León-Castillo et al. 2020 custom POLE framework · vleon-castillo-2020-custom-framework-v1
0%
complete
Re-run interpretation
Re-run this variant?
This variant was interpreted by pipeline
7.1.5.
The current version is
8.0.0.
Re-running queues a fresh interpretation; the result replaces this page when complete.
Re-runs are not free. Blank = draws on today's public interpretation allowance.
With a code = uses 1 of that code's uses.
POLE encodes the catalytic subunit of DNA polymerase epsilon, the enzyme that replicates the leading strand of DNA during cell division and participates in DNA repair. It contains a proofreading domain that corrects replication errors, keeping the accumulation of mutations in check. Germline mutations in POLE cause polyposis and predispose to colorectal cancer, and are also linked to a rare syndrome of facial dysmorphism, immunodeficiency, livedo, and short stature. Somatic mutations, particularly in the proofreading domain, occur in colorectal and endometrial cancers, where they drive an ultra-mutated tumor phenotype and are associated with better responses to immune checkpoint inhibitors.
This variant
POLE encodes the catalytic subunit of DNA polymerase epsilon, and germline loss-of-function mutations in it cause polyposis and predispose to colorectal cancer. This frameshift, predicted to trigger nonsense-mediated decay and to remove the C-terminal domains needed for holoenzyme assembly, would eliminate this essential replication and repair function, consistent with the gene's established loss-of-function disease mechanism and supporting a Likely Pathogenic classification.
Transcript
NM_006231.4
HGVS · transcript:coding
NM_006231.4:c.4519_4520del
GRCh38
chr12:132643254 CTG>C
GRCh37
chr12:133219840 CTG>C
No official ClinGen POLE VCEP specification exists, so standard ACMG/AMP 2015 combination logic was applied; only PVS1 (Very Strong) and PM2 (Moderate) are met.
Classification rationale
PVS1PM2Likely Pathogenic
POLE c.4519_4520delframeshift · exon 35
PVS1 (Very Strong): 2-bp frameshift (p.Gln1507AlafsTer37) in exon 35 of 49 predicted to trigger nonsense-mediated decay, removing essential C-terminal domains. PM2 (Moderate): highest observed allele frequency 0.000176%, far below the <0.1% rarity threshold. PVS1 (Very Strong) plus PM2 (Moderate) under standard ACMG/AMP 2015 combination rules supports Likely Pathogenic.
PVS1 + PM2→Likely Pathogenic
LYFE Sciences is an AI system, and it can make mistakes. Criteria
may be applied incorrectly, sources may be misread, and a confident-looking
classification can still be wrong. Double-check every criterion and
its underlying evidence before relying on any call.
Gene diagram
· NM_006231.4 · variants mapped to exon structure
POLENM_006231.4
Fetching transcript structure from UCSC…
Exons
—
Transcript span
—
Strand
—
Variants mapped
—
Source
UCSC ncbiRefSeqCurated
All variants in POLE—click a row to locate it on the plot · use the link column to open its page
Protein
Location
Classification
Link
Applied criteria · 2 applied · 16 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
✓
PVS1very strongPathogenic
Met (Very Strong): 2-bp frameshift (p.Gln1507AlafsTer37) in exon 35 of 49 predicted to trigger nonsense-mediated decay.
pvs1_variant_assessment.json: consequence_class='frameshift', variant_bucket='frameshift', protein_1l='NP_006222.2:p.(Q1507Afs*37)', apply_generic_pvs1_framework=true, suggested_default_strength='PVS1', framework_source=PMC6185798.pvs1_gene_context.json: germline_disease_context_found=true, lof_mechanism_supported=true, pvs1_gene_gate='eligible'; no official CSPEC/VCEP POLE PVS1 framework exists so generic fallback per PMC6185798 was used.prefetch.json VariantValidator exon/CDS map: variant falls in coding exon 35 of 49 total exons (c.4445-4551), far from the terminal exon/penultimate-exon 50bp NMD-escape boundary, supporting NMD.
Met (Moderate): highest observed allele frequency 0.000176%, far below the <0.1% rarity threshold.
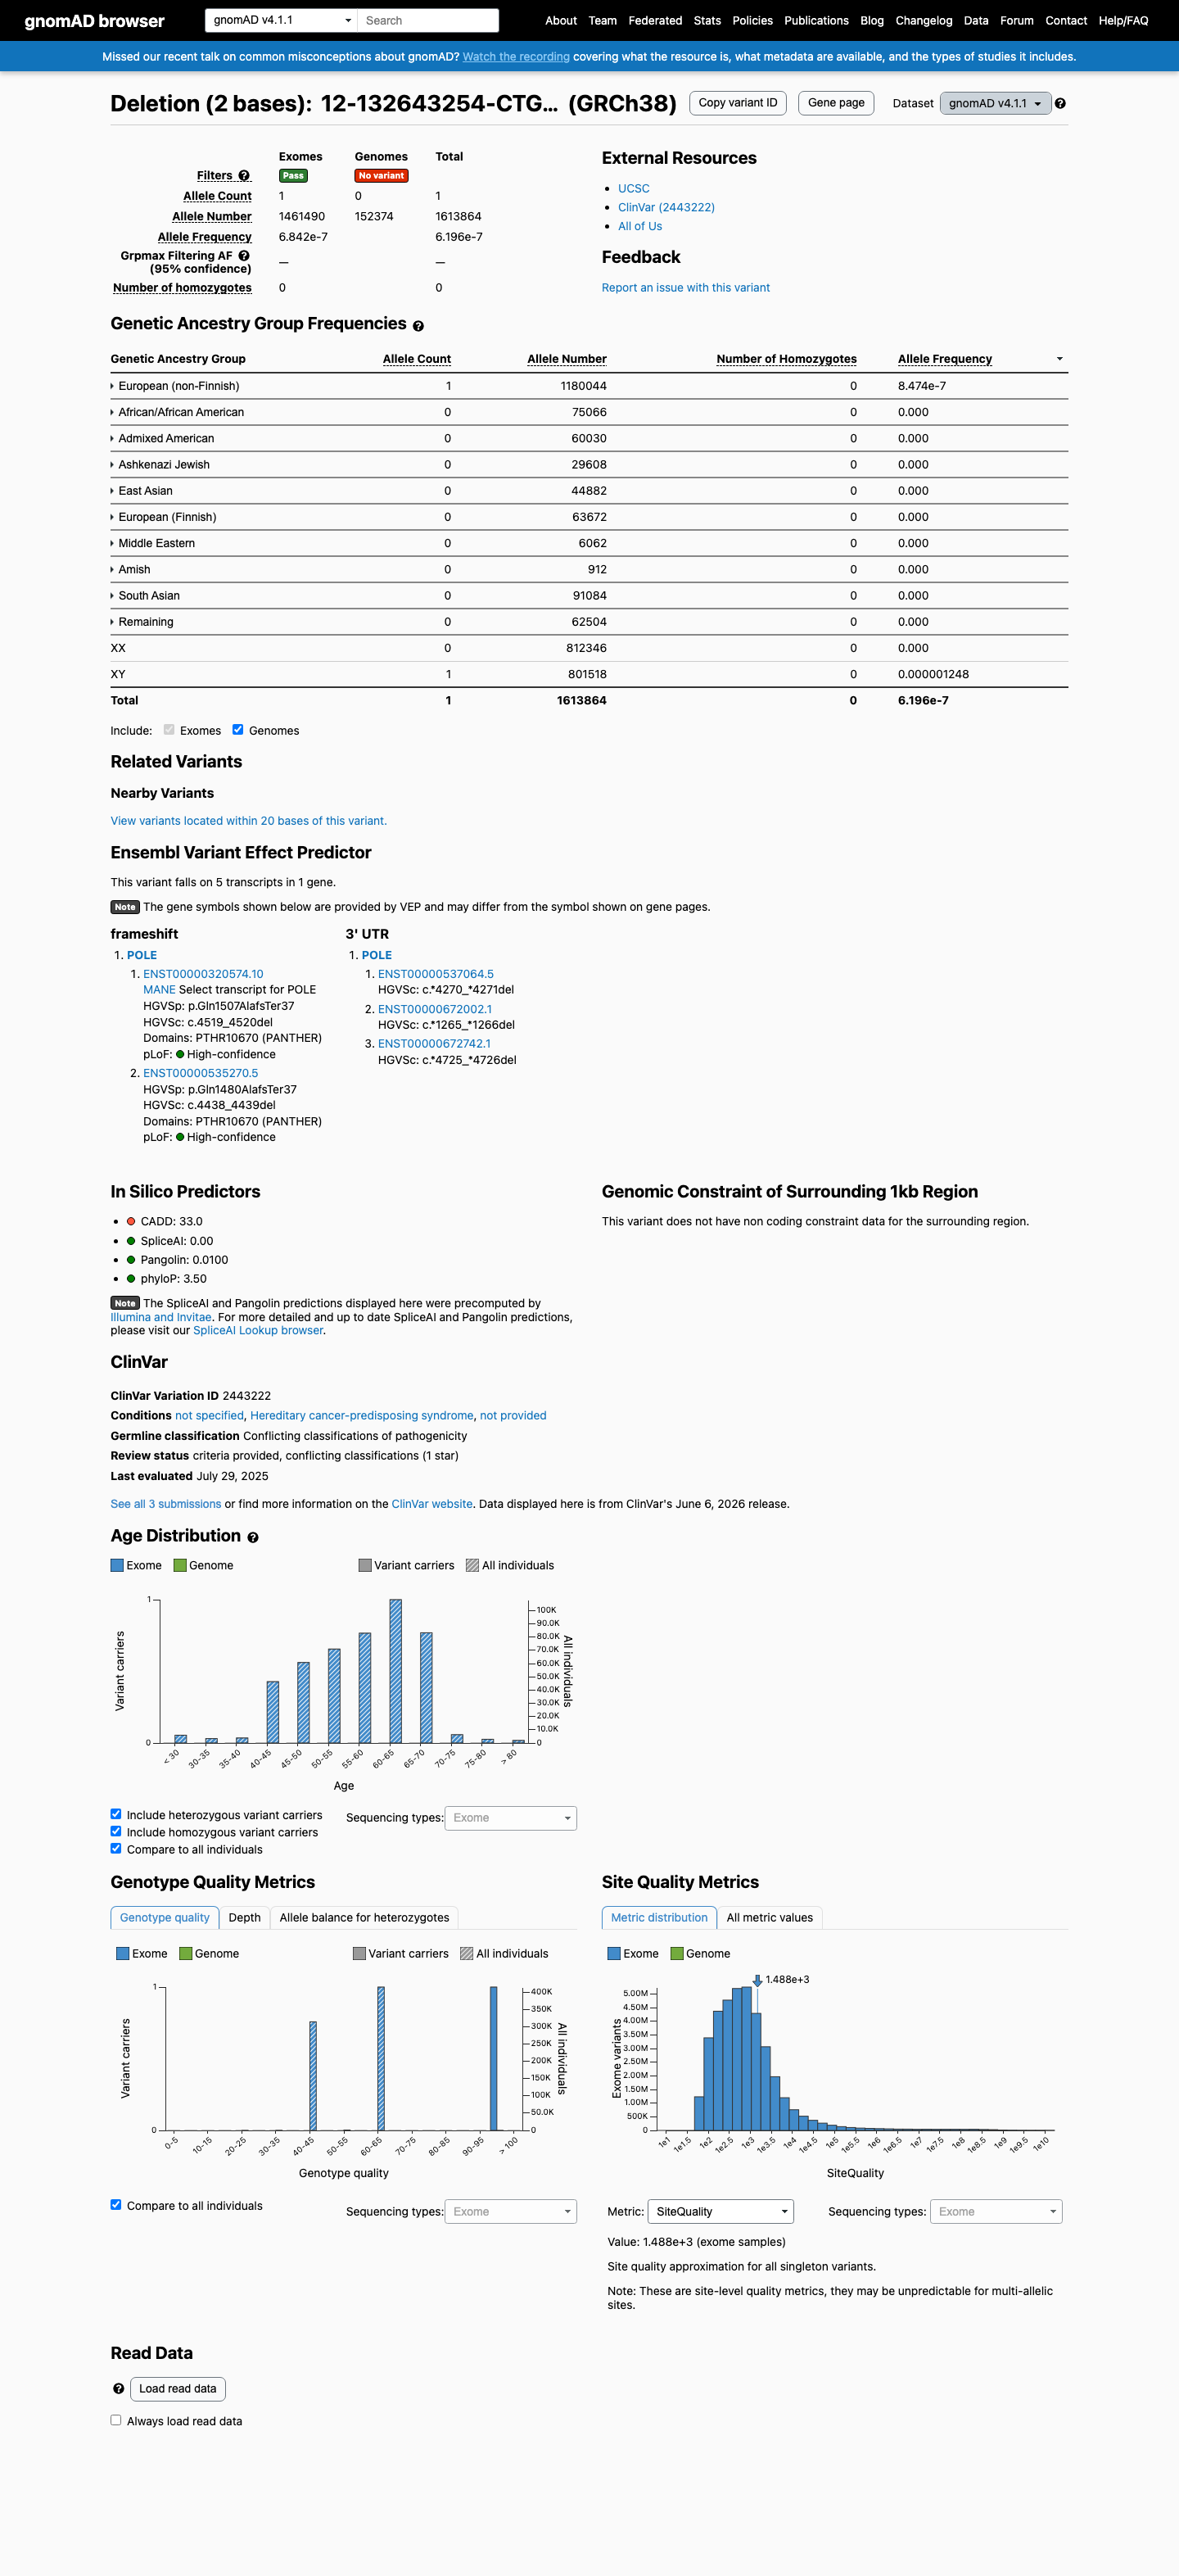
gnomAD v2.1 reports the variant as absent.gnomAD v4.1 reports one allele among 1,613,864 total alleles (overall AF 6.19631e-07; 0 homozygotes); highest reported frequency is 1/568,400 non-Finnish European XY alleles (AF 1.75932e-06).gnomAD-Canada v1.0 reports the variant as absent.
This variant is present in gnomAD v4.1 (AF= 6.19631e-07; MAF= 0.00006%, 1/1613864 alleles, homozygotes = 0) and has highest observed frequency in the European (non-Finnish) population (AF= 8.47426e-07; MAF= 0.00008%, 1/1180044 alleles, homozygotes = 0).
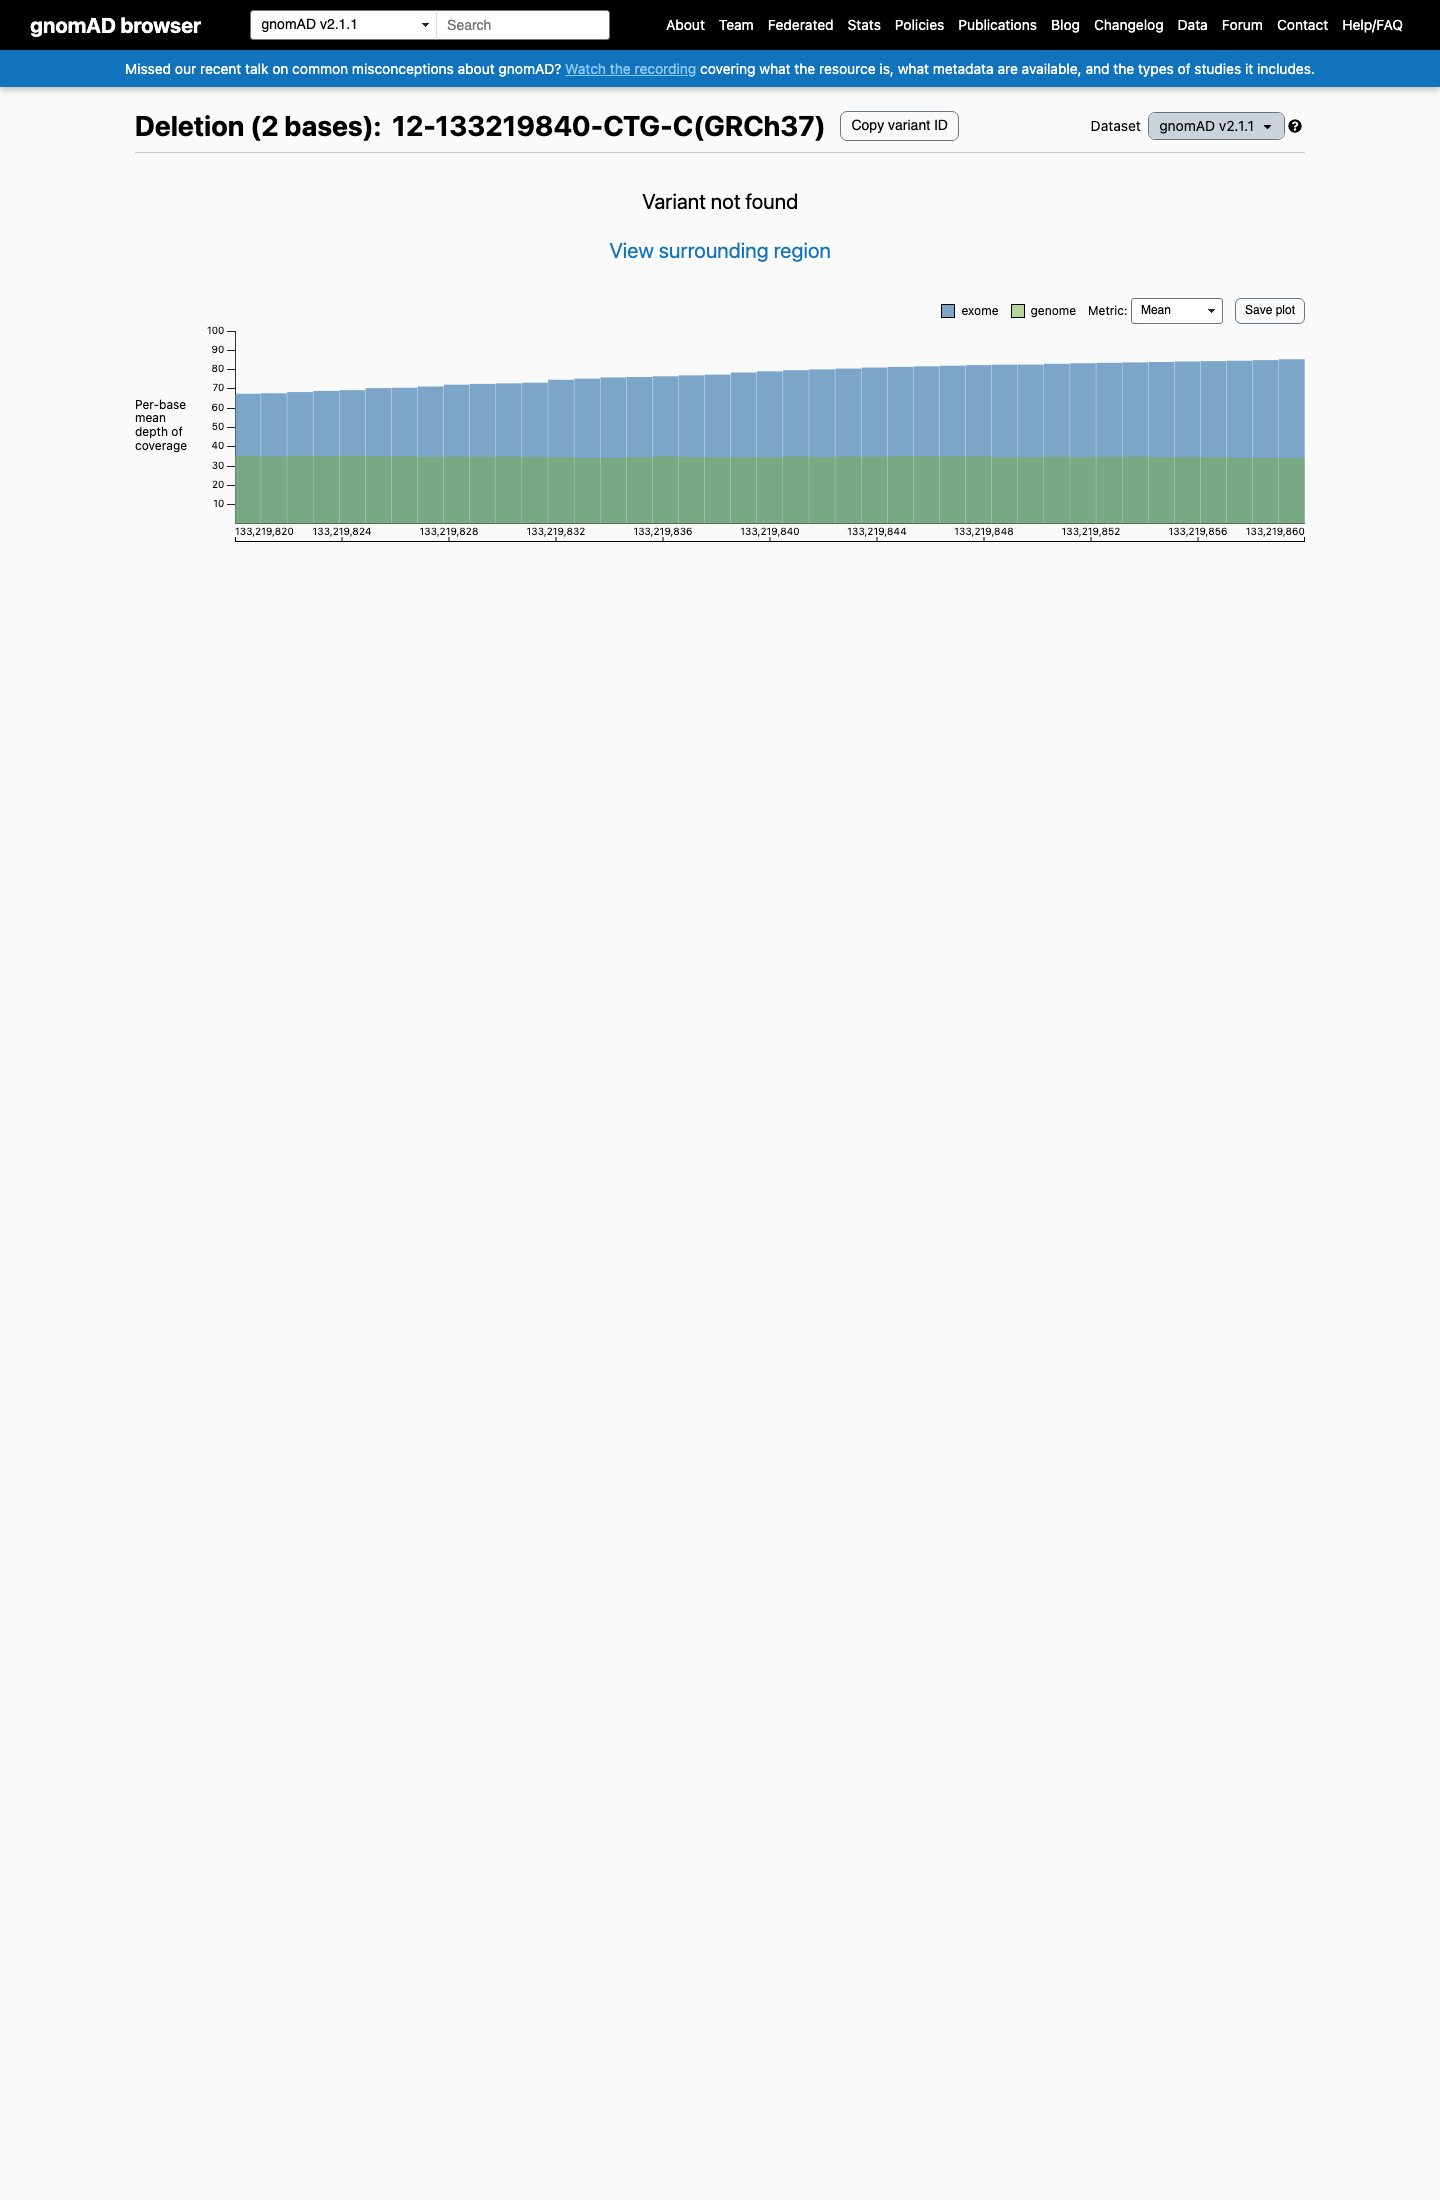
v2.1
Absent from gnomAD v2.1.
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
6.2e-05%
· 1 / 1,613,864
0 hom
European (non-Finnish)
1 / 1,180,044
8.5e-05%
+ 9 not observed (Remaining individuals, Admixed American, European (Finnish), Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American)
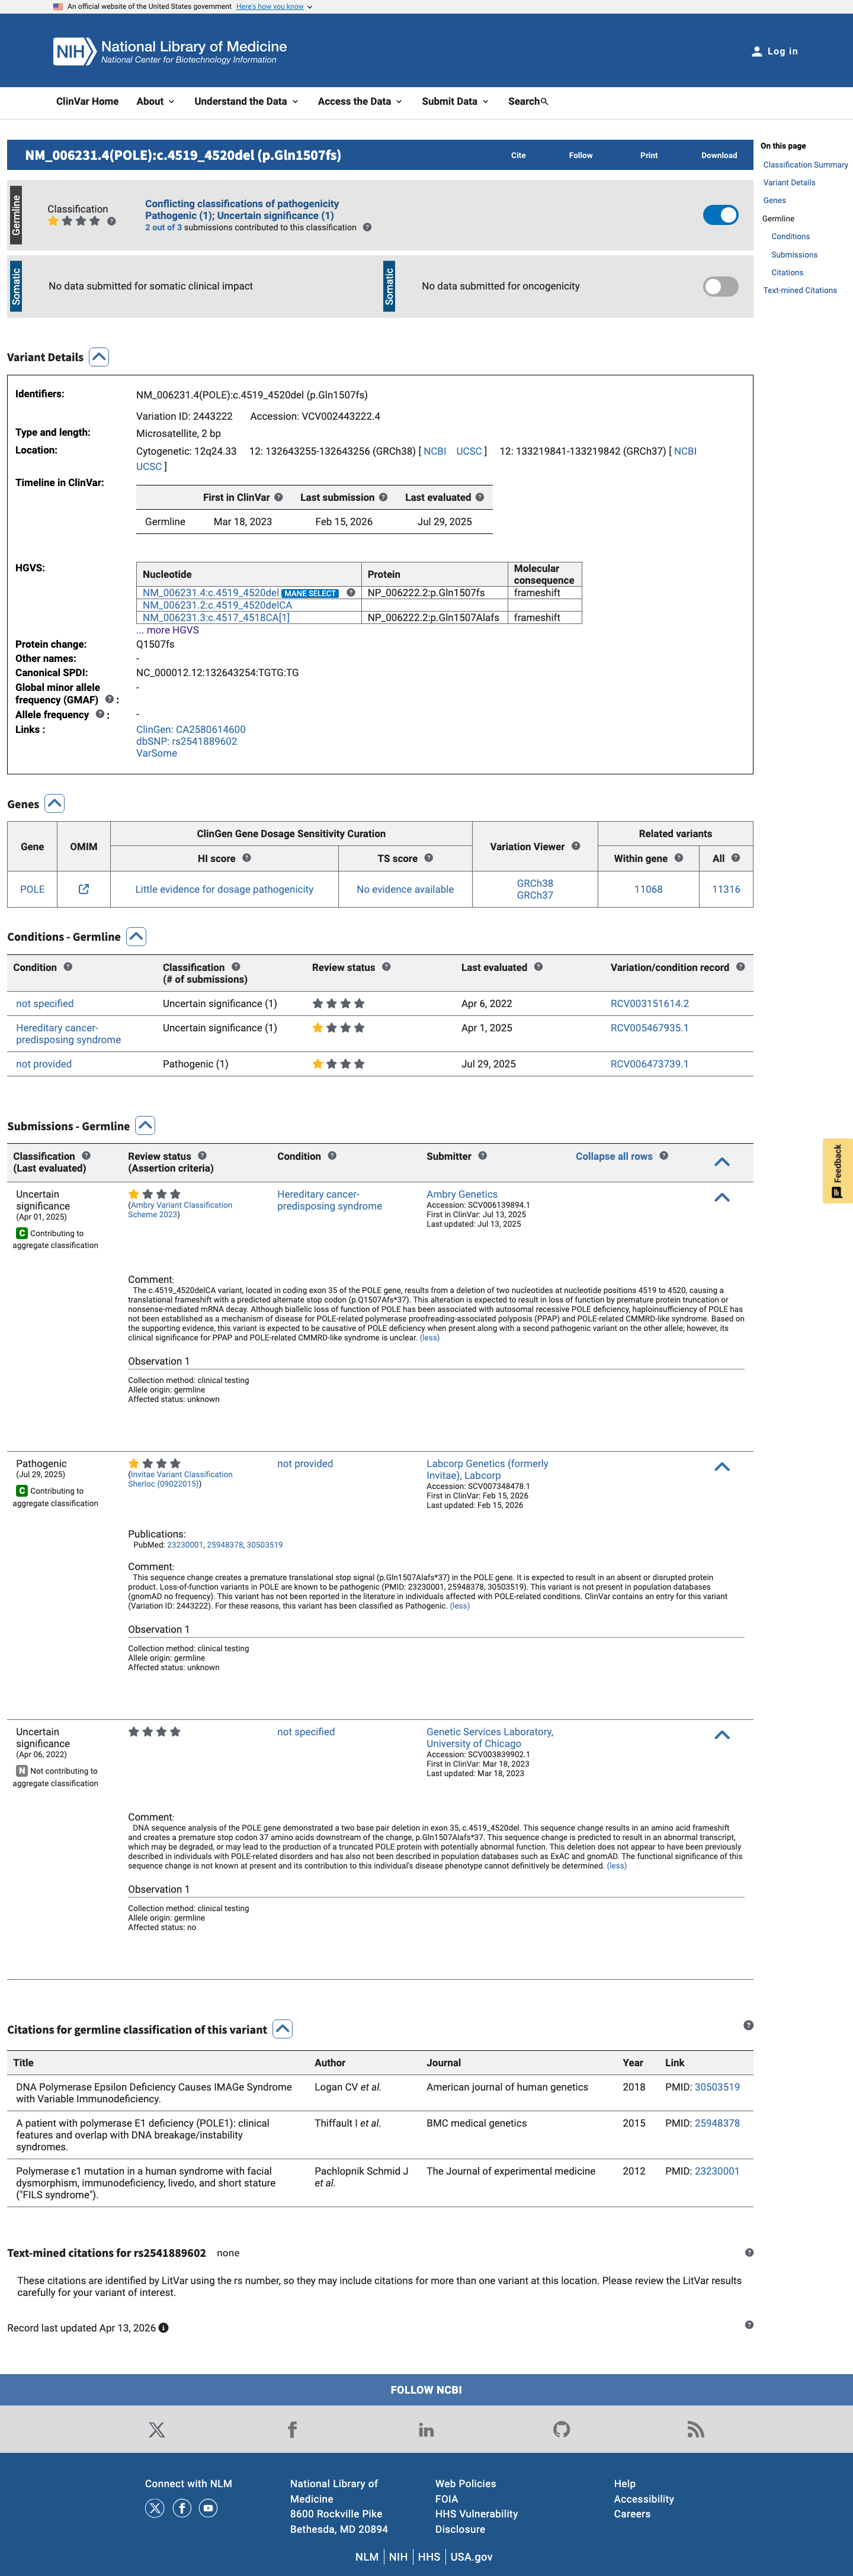
This variant has been reported in ClinVar as Uncertain significance (2 clinical laboratories) and as Pathogenic (1 clinical laboratory). (ClinVarID = 2443222)
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. POLE, the catalytic subunit of DNA polymerase epsilon, is an enzyme involved in DNA replication and repair. Select POLE mutations lead to ultra-high m
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 2 further PMIDs triaged but not cited — see Sources & references.
Rule & framework references · cited for criterion definitions, not variant evidence
23230001 ↗Polymerase ε1 mutation in a human syndrome with facial dysmorphism, immunodeficiency, livedo, and short stature ("FILS syndrome").
Triaged references · 2 PMIDs not cited in assessment
25948378 ↗A patient with polymerase E1 deficiency (POLE1): clinical features and overlap with DNA breakage/instability syndromes.CLINVAR
25394175 ↗A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment.CLINVAR