Back
NM_001903.5:c.710A>G
p.Tyr237Cys · CTNNA1
ACMG/AMP
0%
complete
Final classification
VUS
PM2BP4
CTNNA1
c.710A>G
p.Tyr237Cys
missense · exon 6

CTNNA1 encodes a member of the catenin protein family that plays a central role in cell adhesion by linking cadherins on the cell surface to the actin cytoskeleton; its mechanosensing activity lets cells respond to physical tension and reorganize these connections. Mutations in this gene cause butterfly-shaped pigment dystrophy of the eye. CTNNA1 also acts as a tumor suppressor: its inactivation promotes cellular invasiveness and metastasis, and reduced expression has been observed in several cancers, including bladder cancer, acute myeloid leukemia, and breast cancer.

This variant

CTNNA1 encodes an adhesion-related catenin that links cadherins to the actin cytoskeleton and has tumor-suppressor functions, with disease relevance also reported for butterfly-shaped pigment dystrophy of the eye.

Transcript
NM_001903.5
HGVS · transcript:coding
NM_001903.5:c.710A>G
GRCh38
chr5:138824651 A>G
GRCh37
chr5:138160340 A>G
VUS: PM2 (supporting) and BP4 (supporting) do not satisfy any generic ACMG/AMP pathogenic or benign classification combination.
Classification rationale
PM2 BP4 VUS
CTNNA1 c.710A>G missense · exon 6

PM2 supporting: gnomAD v4.1 allele frequency is below the generic supporting threshold. BP4 supporting: REVEL 0.278 meets the generic supporting benign threshold. VUS: PM2 supporting plus BP4 supporting does not meet a generic ACMG/AMP classification threshold.

PM2 + BP4 VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_001903.5 · variants mapped to exon structure
CTNNA1 NM_001903.5
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 22 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
PM2 supporting Pathogenic
Met at supporting strength: gnomAD v4.1 AF 4.83236e-05 is below the supplied PM2 threshold of 0.0001.
No governing CTNNA1-specific VCEP or gene-specific framework was available; the supplied generic PM2 supporting calibration was applied.gnomAD v4.1 reports AF 4.83236e-05 (78/1,614,118 alleles) and 0 homozygotes.gnomAD v2.1 reports AF 8.75065e-05 (22/251,410 alleles) and 0 homozygotes.
BP4 supporting Benign
Met, supporting: REVEL 0.278 meets the generic BP4 supporting threshold of <=0.29 from the ClinGen SVI calibration.
Variant consequence is missense: NM_001903.5:c.710A>G, NP_001894.2:p.(Tyr237Cys).REVEL score is 0.278; the supplied generic ClinGen SVI BP4 supporting threshold is <=0.29, from Pejaver et al. 2022 (PMID:36413997).BayesDel score is -0.156098, but no generic ACMG-strength calibration is established for BayesDel, so it is not available for BP4 unless a governing VCEP specifies a cutoff.
Assessed · not applied · 7 not met · 15 not assessed
Pathogenic
PS1 Not assessed: no established pathogenic variant with the same Tyr237Cys amino-acid change was identified in the available evidence.
PS2 Not assessed: no confirmed de novo occurrence or parental genotypes are documented for CTNNA1 c.710A>G.
PS3 Not assessed: no validated variant-specific functional assay or abnormal experimental result was available for CTNNA1 p.Tyr237Cys.
PS4 Not assessed: no variant-specific case-control enrichment, prevalence comparison, or affected-case series was identified for CTNNA1 c.710A>G.
PM1 Not met: residue Tyr237 is not in a documented critical domain or statistically significant hotspot, and no gene-specific domain table applies.
PM3 Not assessed: no affected-proband observation, pathogenic variant in trans, or phase evidence is documented for this CTNNA1 variant.
PM5 Not assessed: no pathogenic or likely pathogenic alternate missense variant at CTNNA1 residue 237 was identified for comparison.
PM6 Not assessed: no presumed de novo occurrence with incomplete parental testing is documented for CTNNA1 c.710A>G.
PP1 Not assessed: zero informative affected-relative or meiosis data are documented for CTNNA1 c.710A>G.
PP2 Not assessed: no calibrated CTNNA1 missense-spectrum evidence demonstrates high benign and low pathogenic missense rates for PP2.
PP3 Not met: REVEL 0.278 is below the PP3 supporting threshold of 0.644, and no calibrated BayesDel cutoff or CTNNA1-specific rule is available.
PP4 Not assessed: no patient phenotype or highly specific disease feature is documented for this exact CTNNA1 variant.
PP5 Not met: ClinVar reports zero expert-panel submissions and no exact-variant Pathogenic or Likely pathogenic expert-panel classification.
Benign
BA1 Not met: maximum reported allele frequency is 0.0003502627, far below the generic BA1 stand-alone threshold of 0.05.
BS1 Not met: maximum reported allele frequency is 0.0003502627, below the supplied generic BS1 threshold of 0.01.
BS2 Not met: no homozygotes were observed in gnomAD v2.1, v4.1, or the available non-cancer population subsets.
BS3 Not assessed: no validated variant-specific functional assay or preserved-function result was available for CTNNA1 p.Tyr237Cys.
BS4 Not assessed: no affected relatives lacking CTNNA1 c.710A>G or other informative non-segregation data are documented.
BP1 Not assessed: CTNNA1 loss-of-function evidence is present, but a primarily truncating disease spectrum required for BP1 is not established.
BP2 Not assessed: no individual-level cis/trans phase or pathogenic-variant pairing is documented for this CTNNA1 variant.
BP5 Not assessed: no alternative pathogenic molecular cause is documented in an affected individual carrying this CTNNA1 variant.
BP6 Not met: the only Benign ClinVar assertion is a single-submitter laboratory result, not an exact-variant expert-panel classification.
N/A · 4 PVS1 · PM4 · BP3 · BP7
Research & evidence
Population frequency
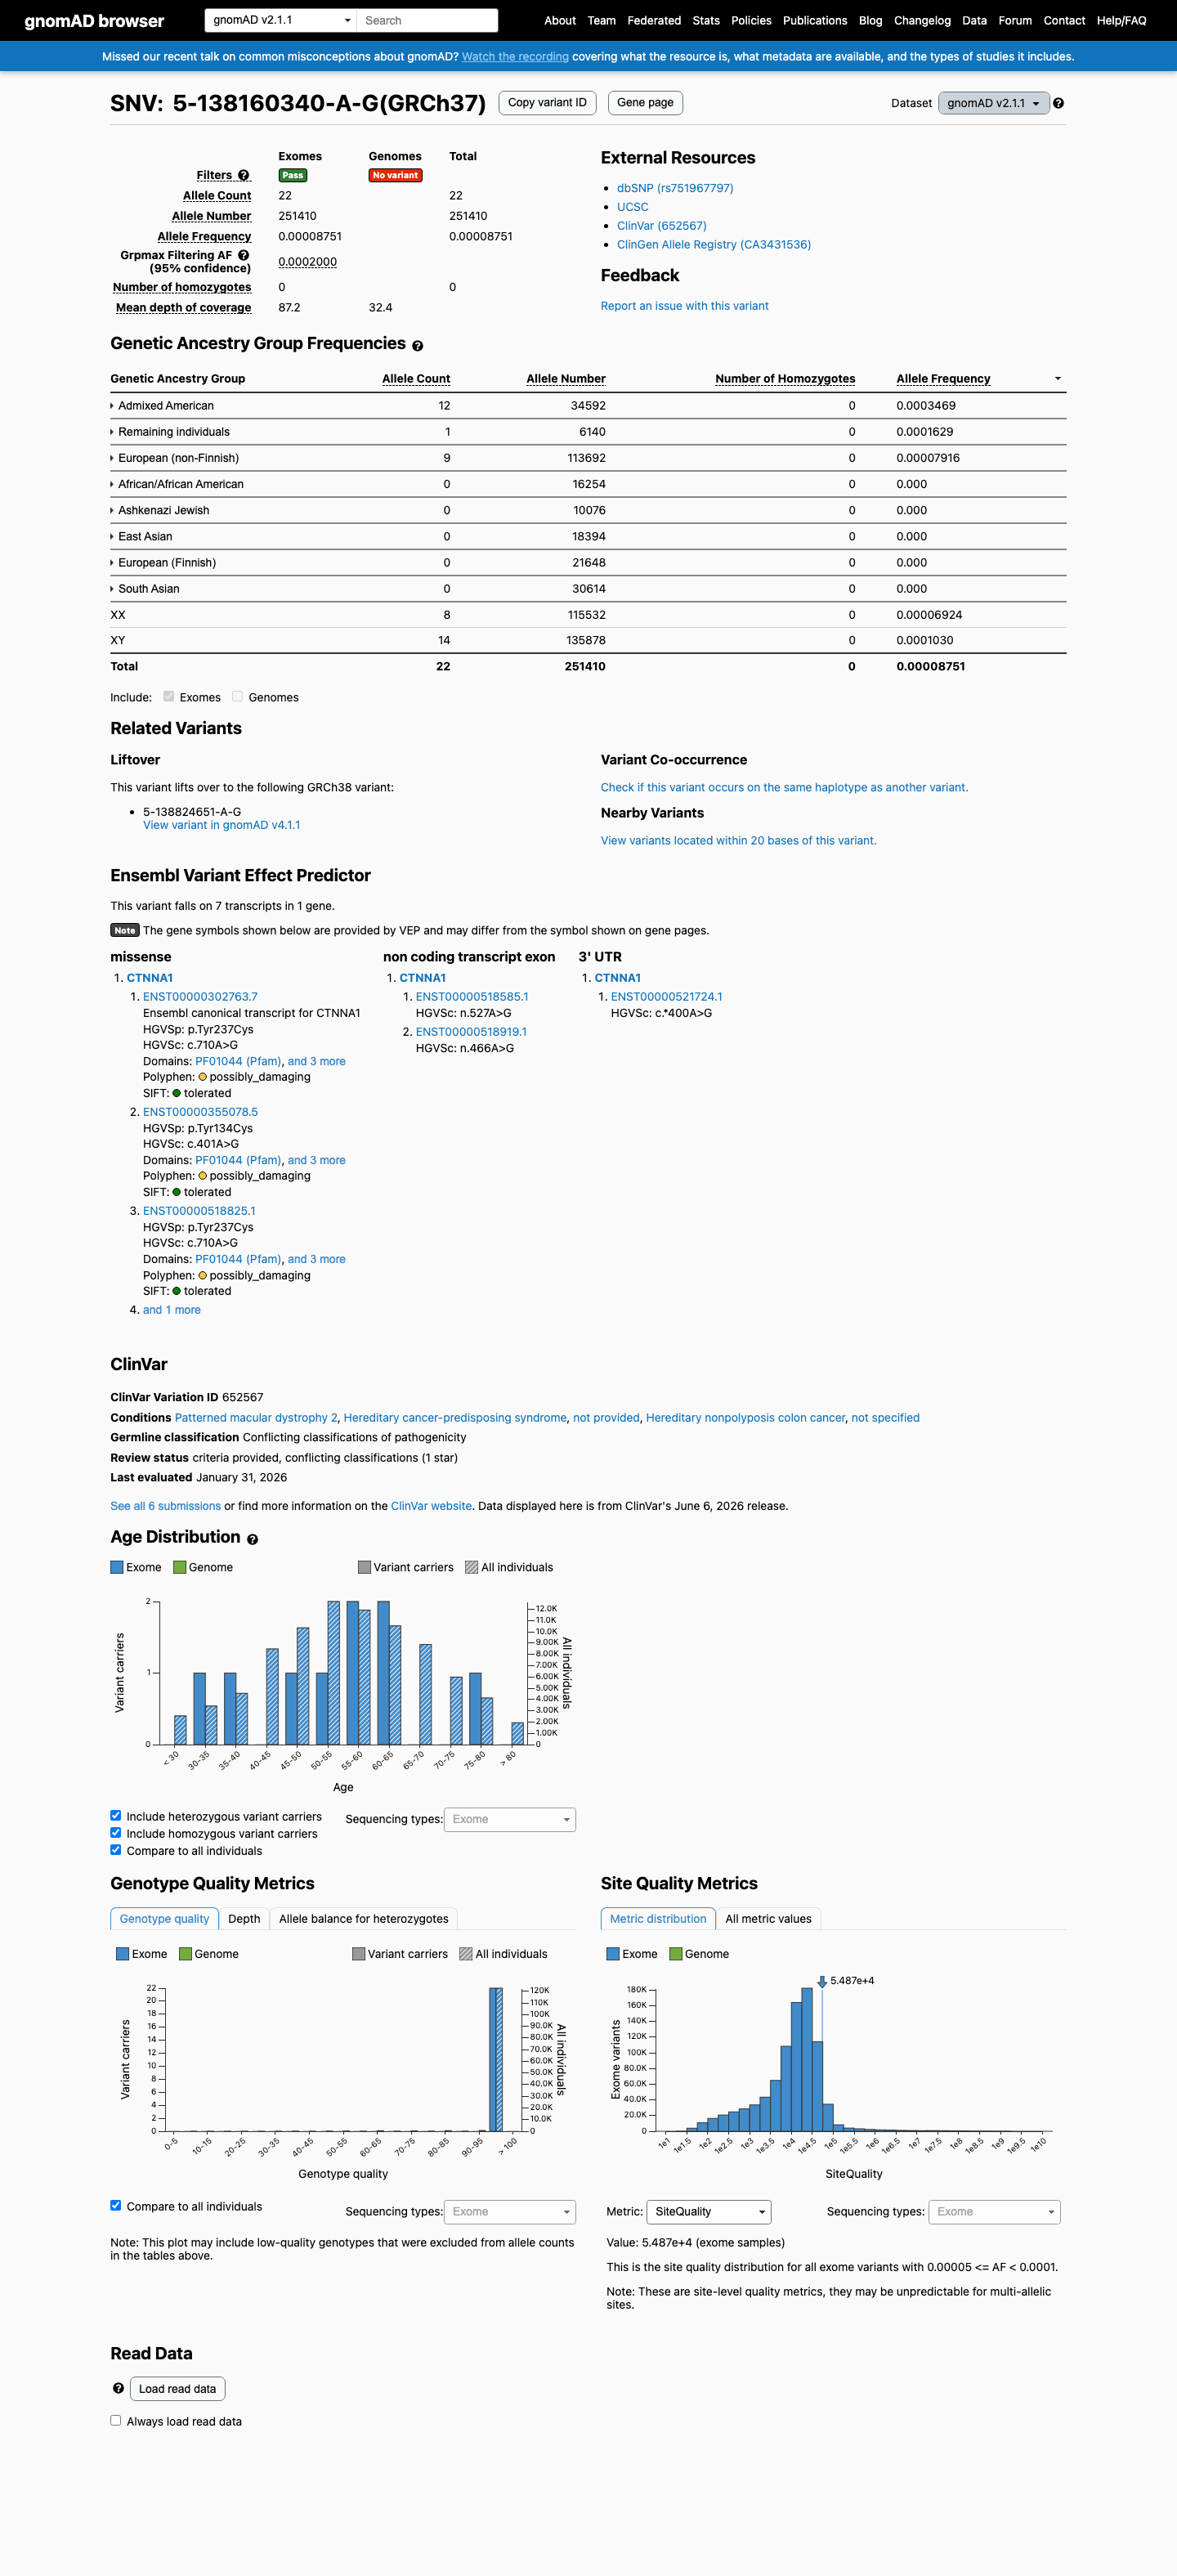
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
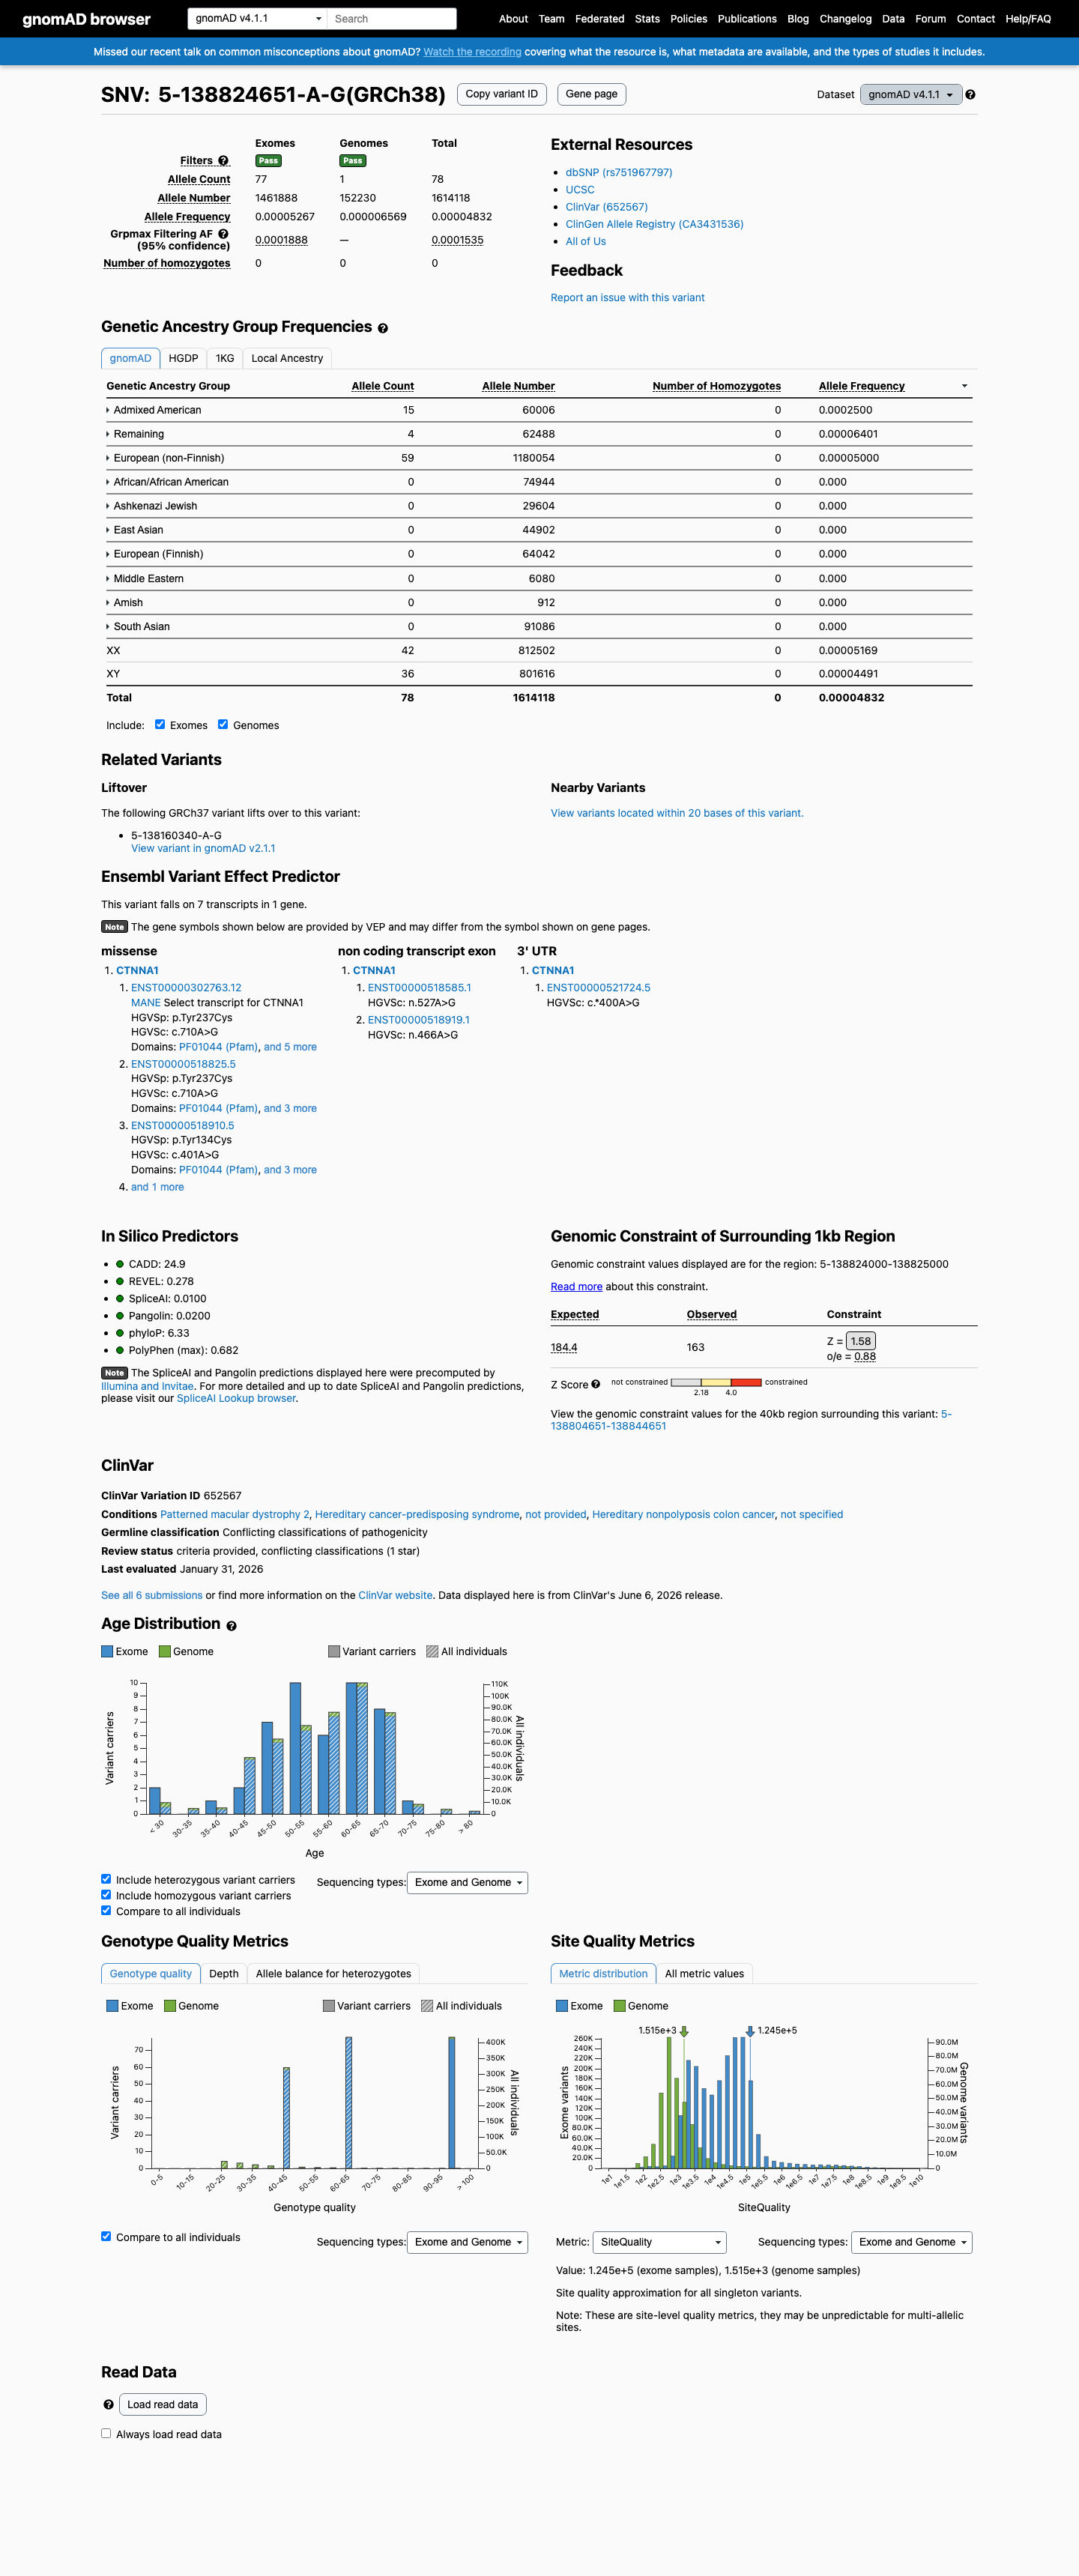
This variant is present in gnomAD v4.1 (AF= 4.83236e-05; MAF= 0.00483%, 78/1614118 alleles, homozygotes = 0) and has highest observed frequency in the Admixed American population (AF= 0.000249975; MAF= 0.02500%, 15/60006 alleles, homozygotes = 0); grpmax FAF= 0.00015354.
v2.1
This variant is present in gnomAD v2.1 (AF= 8.75065e-05; MAF= 0.00875%, 22/251410 alleles, homozygotes = 0) and has highest observed frequency in the Admixed American population (AF= 0.000346901; MAF= 0.03469%, 12/34592 alleles, homozygotes = 0); grpmax FAF= 0.00020003.
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.0048% · 78 / 1,614,118
0 hom · FAF 0.015%
Admixed American
15 / 60,006
0.025%
Remaining individuals
4 / 62,488
0.0064%
European (non-Finnish)
59 / 1,180,054
0.005%
+ 7 not observed (European (Finnish), Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American)
gnomAD v2.1
0.0088% · 22 / 251,410
0 hom · FAF 0.02%
Admixed American
12 / 34,592
0.035%
Remaining individuals
1 / 6,140
0.016%
European (non-Finnish)
9 / 113,692
0.0079%
+ 5 not observed (African/African American, Ashkenazi Jewish, East Asian, European (Finnish), South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
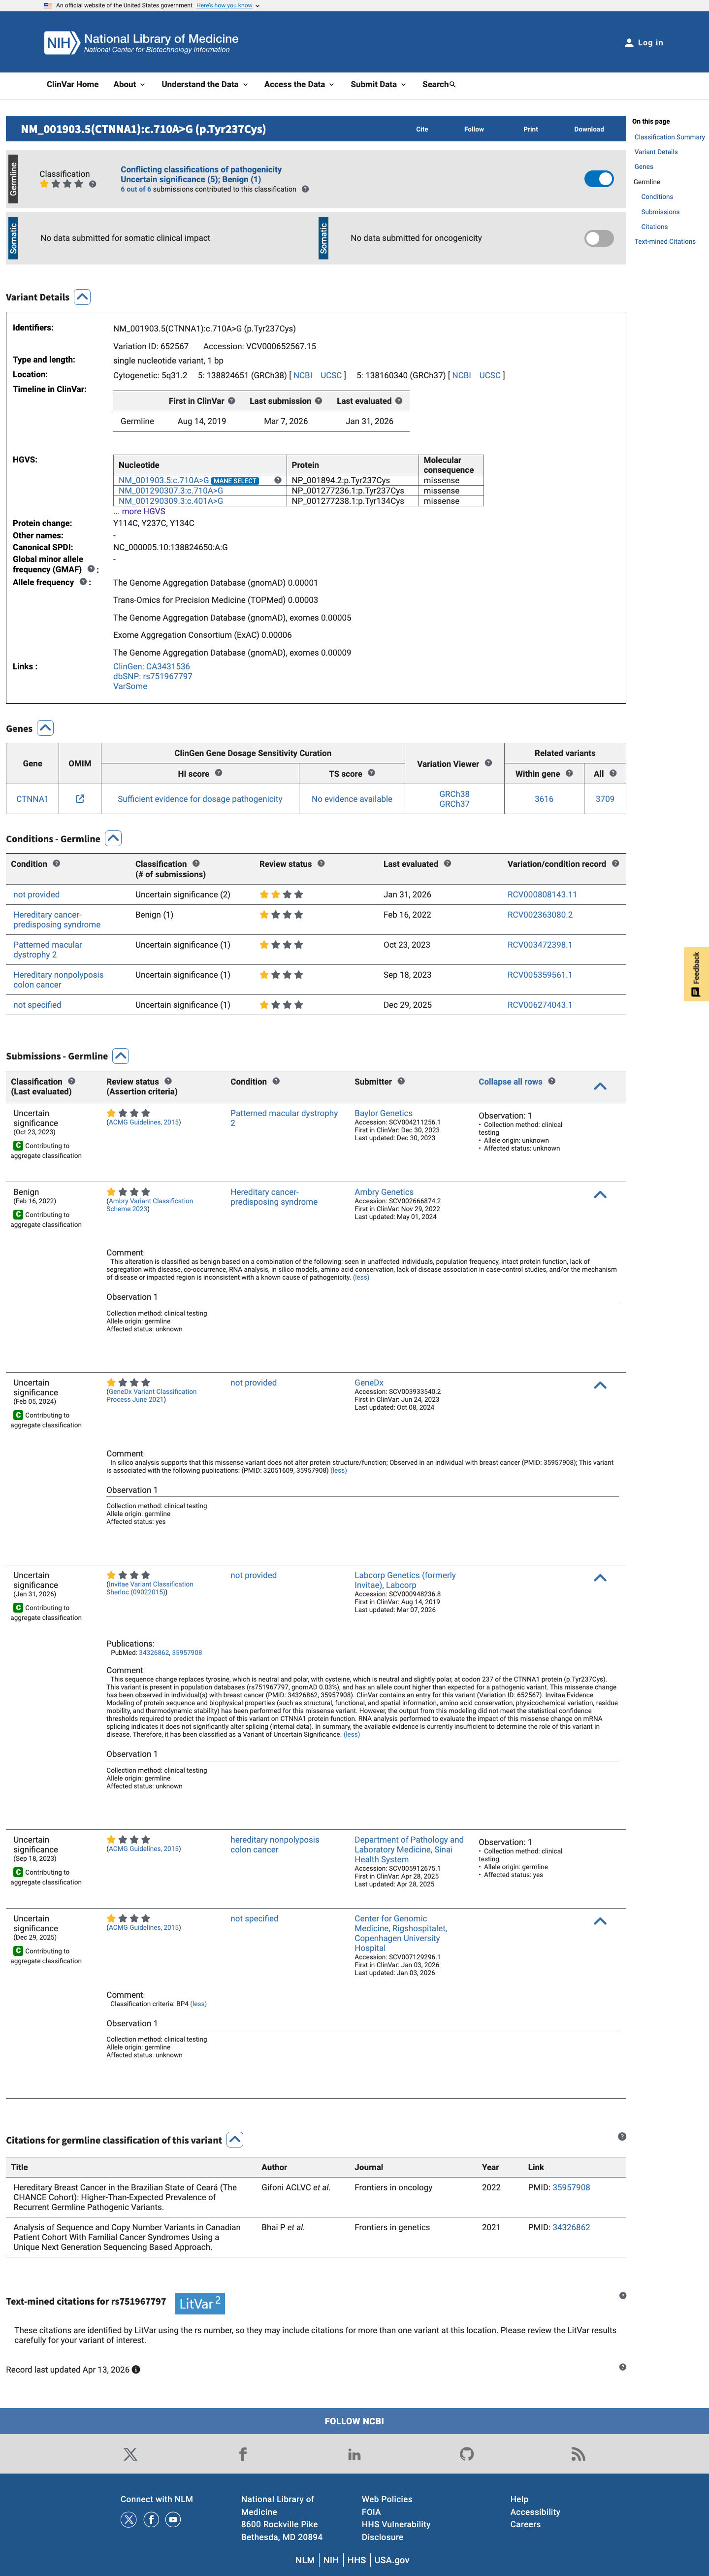
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Uncertain significance (5 clinical laboratories) and as Benign (1 clinical laboratory). (ClinVarID = 652567)
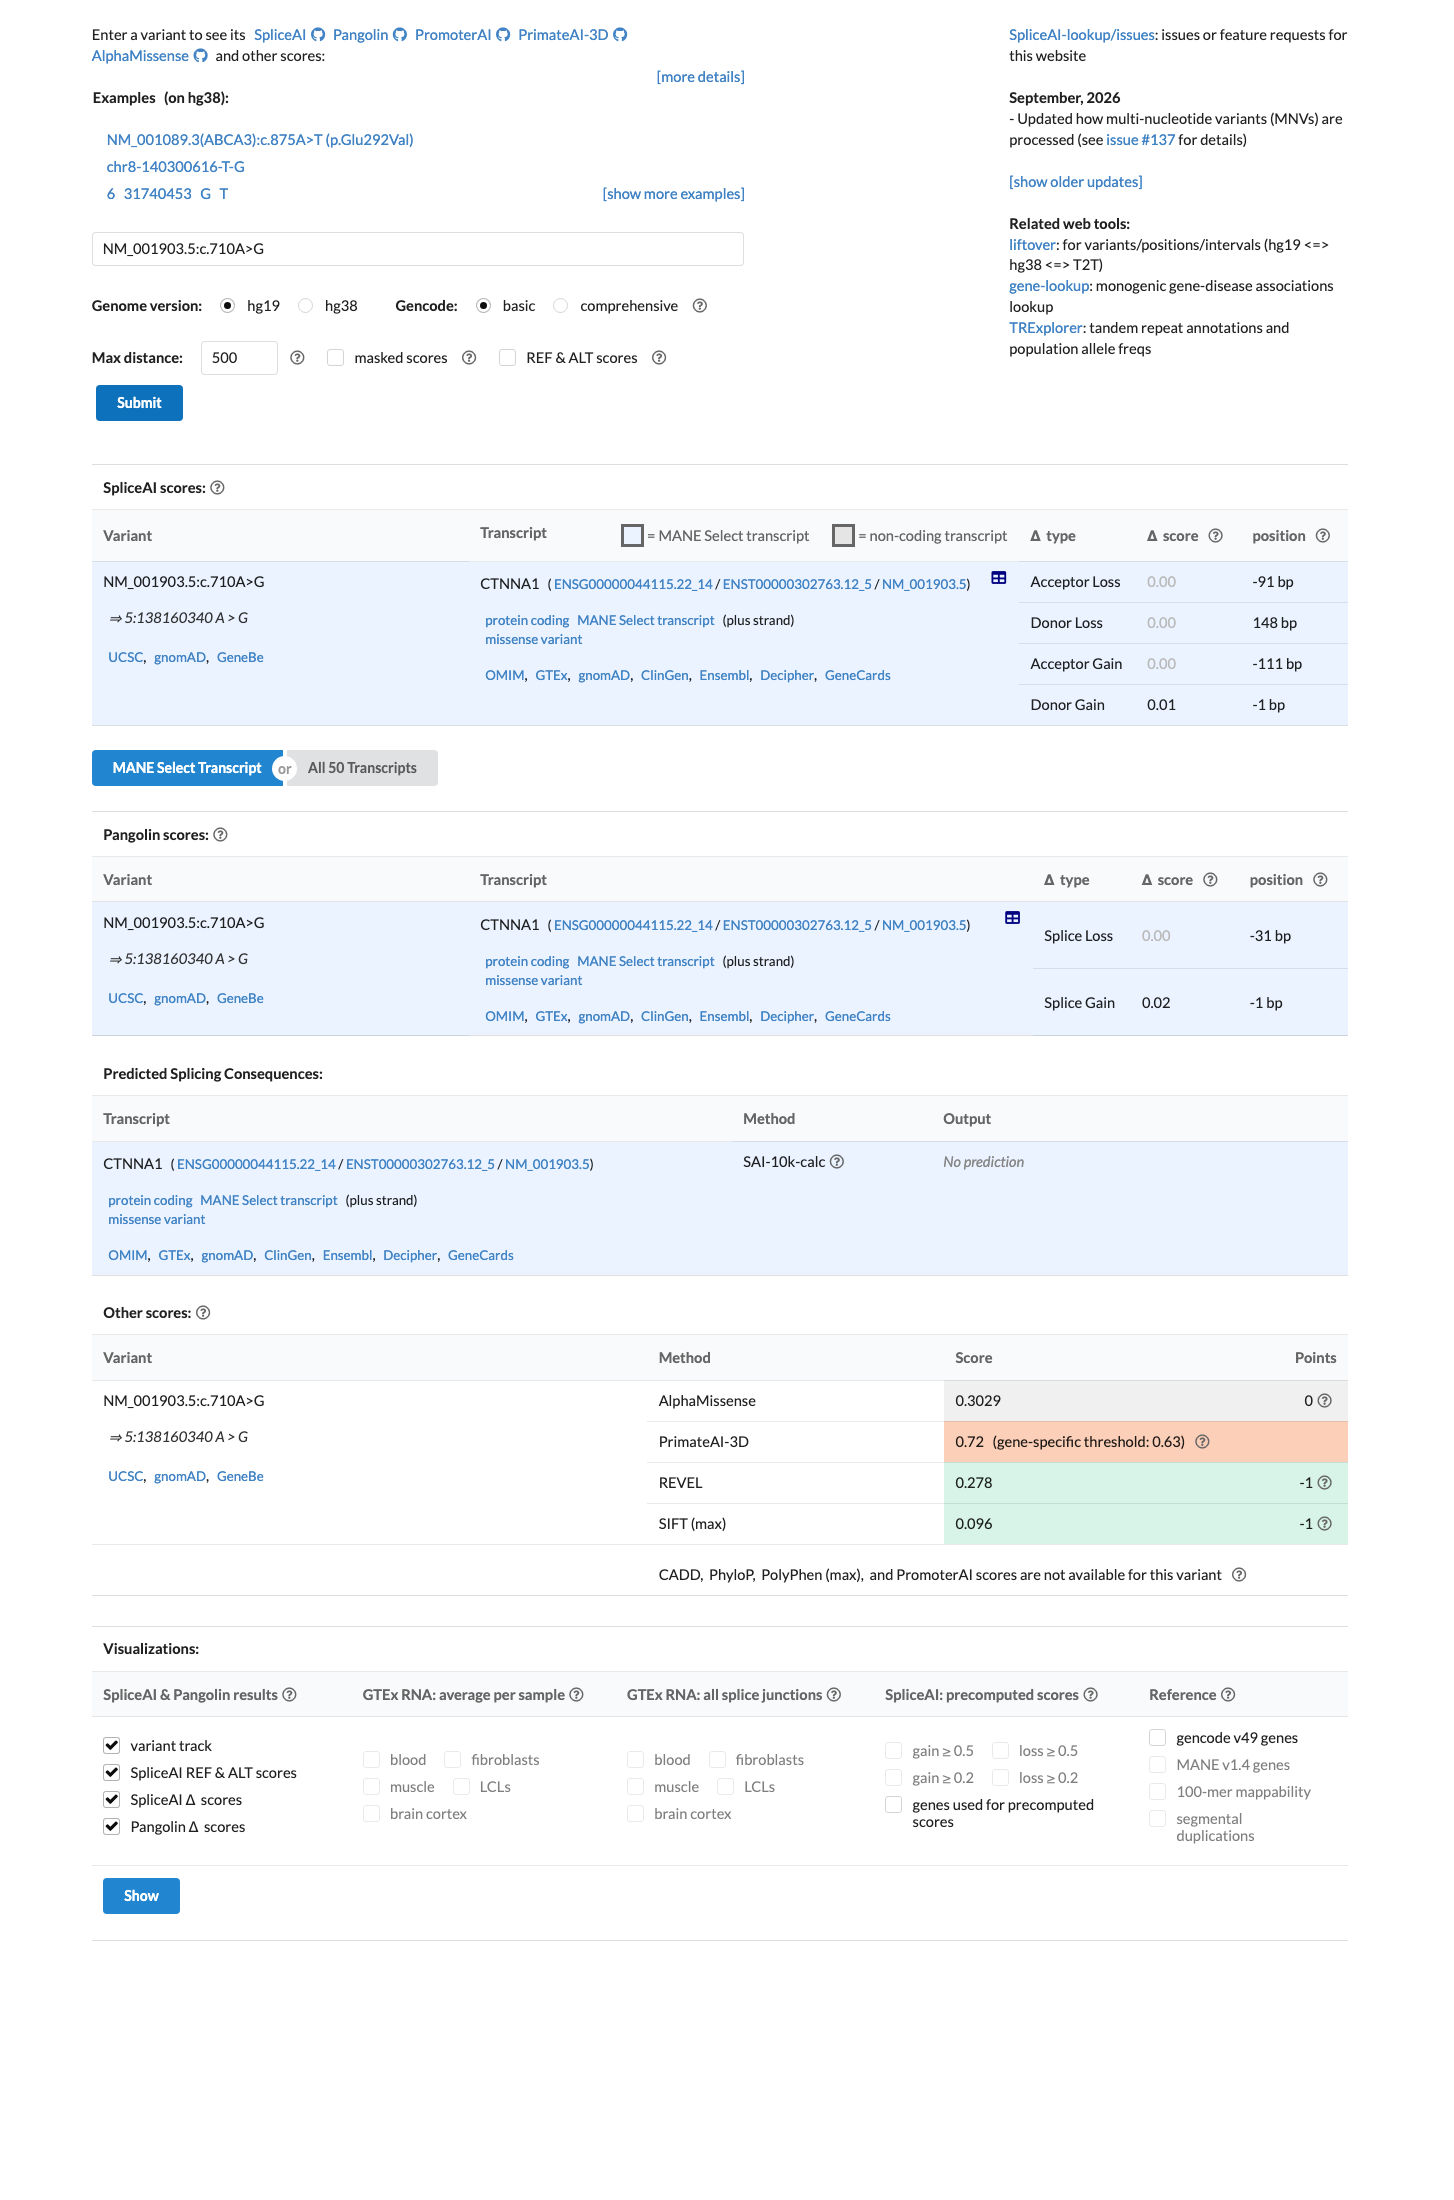
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.01). REVEL score = 0.278. BayesDel score = -0.156098.
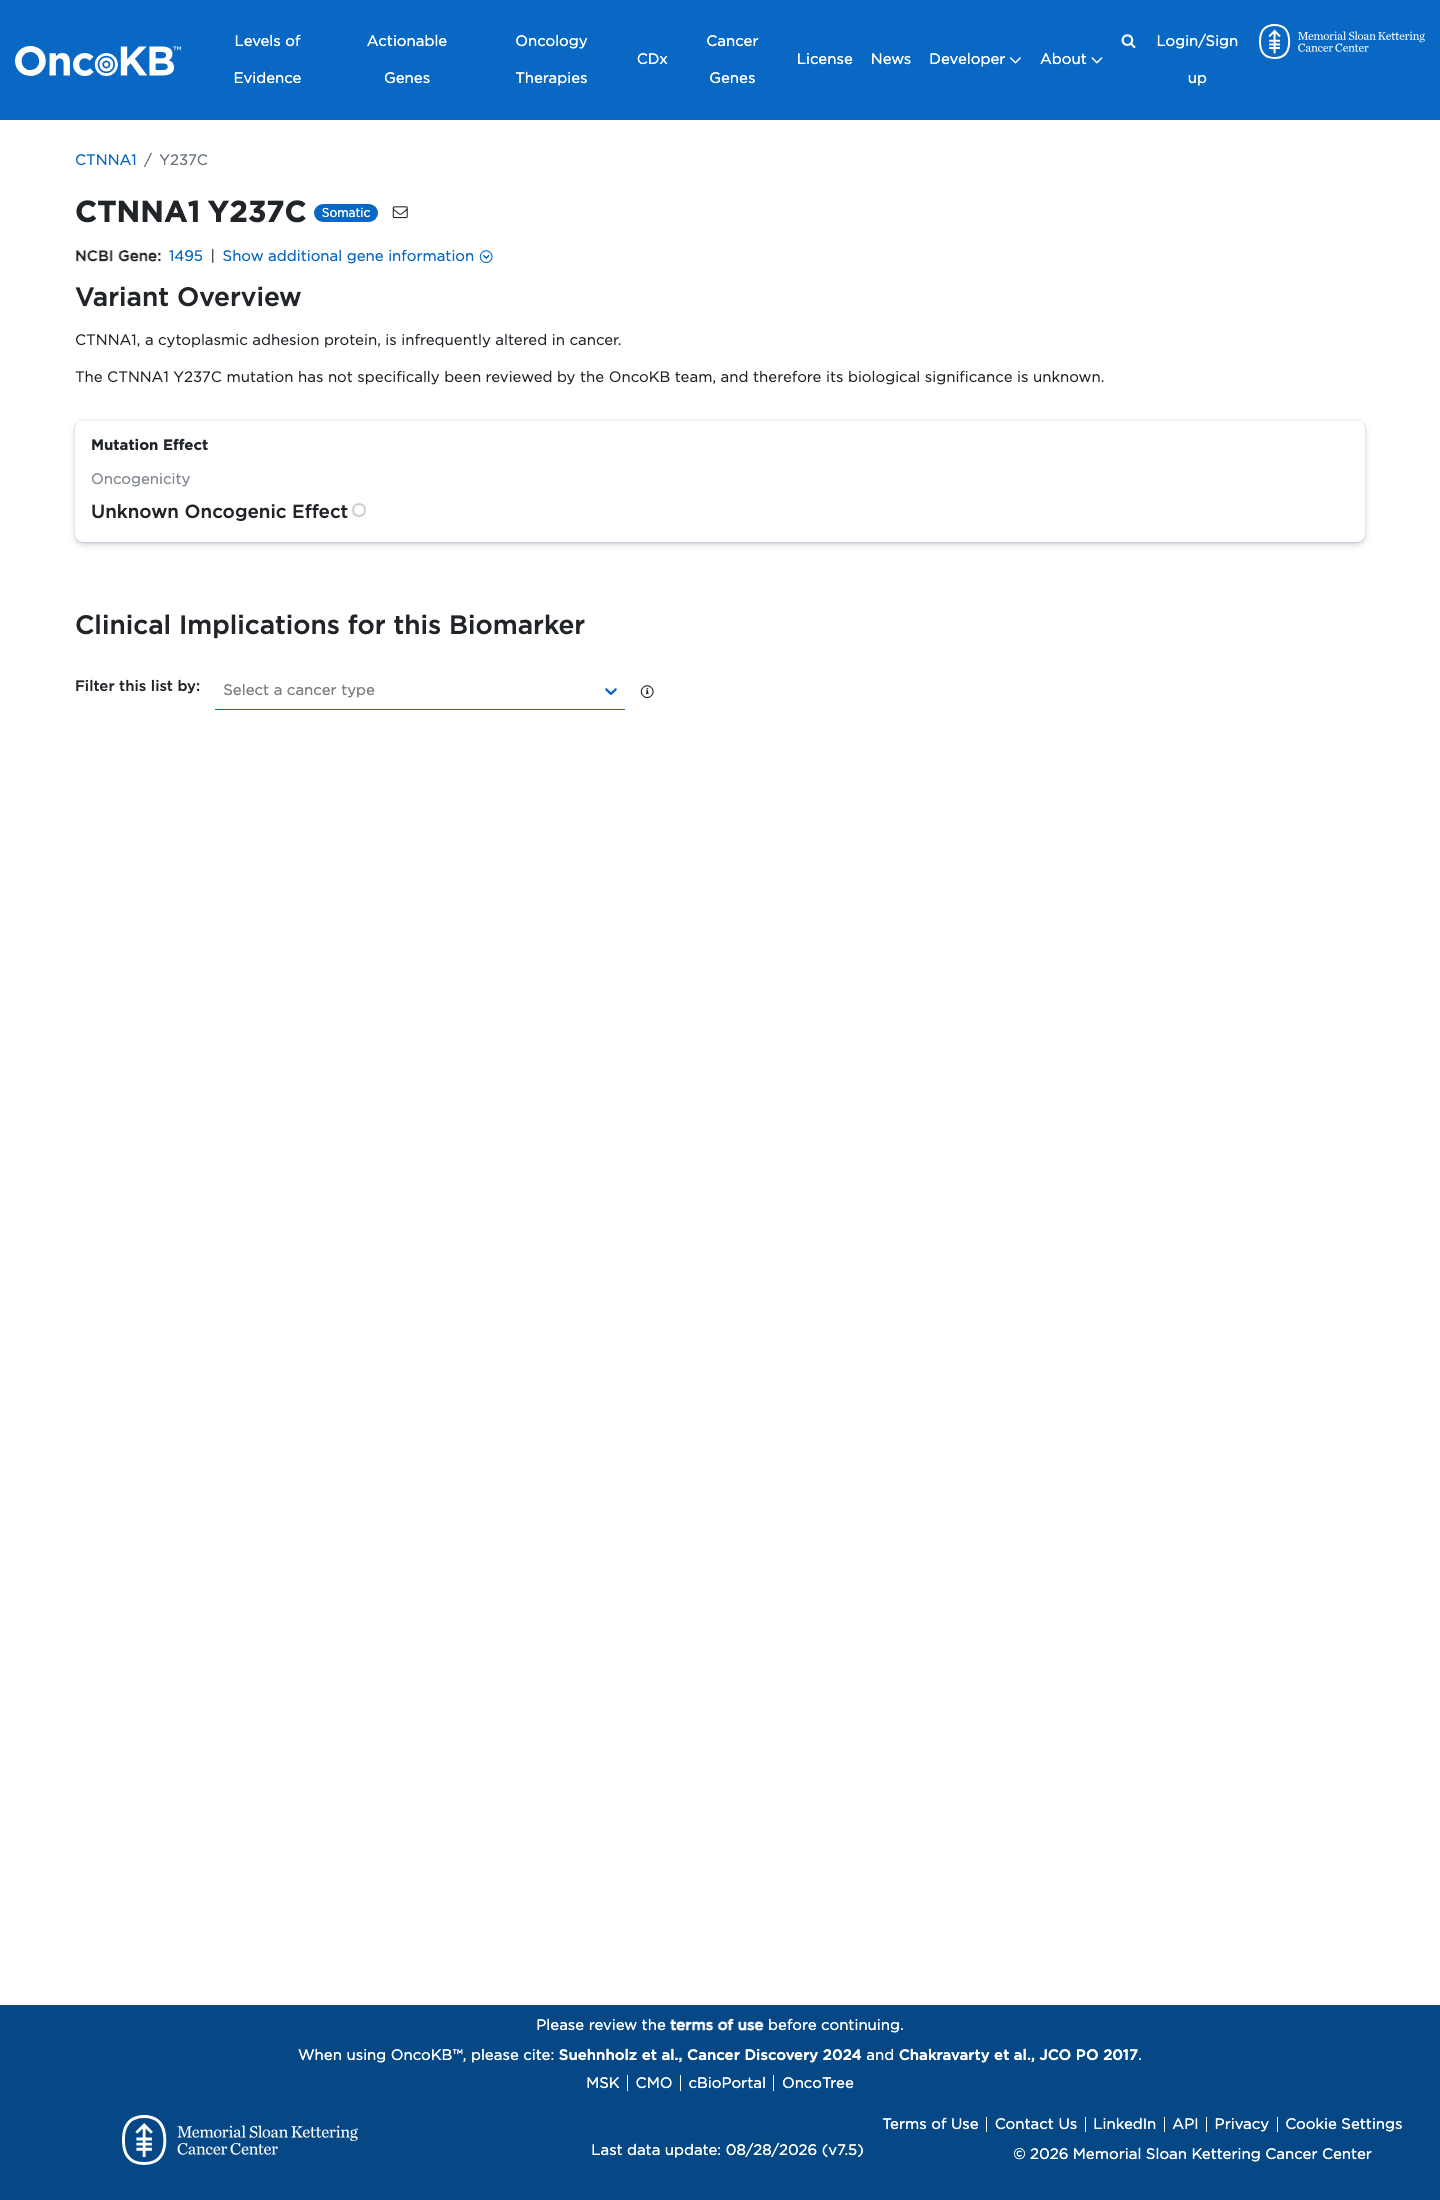
Functional / OncoKB screenshot
Functional Unknown Oncogenic Effect
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. CTNNA1, a cytoplasmic adhesion protein, is infrequently altered in cancer.
OncoKB ↗
COSMIC screenshot
COSMIC
Cancer hotspots screenshot
Cancer hotspots
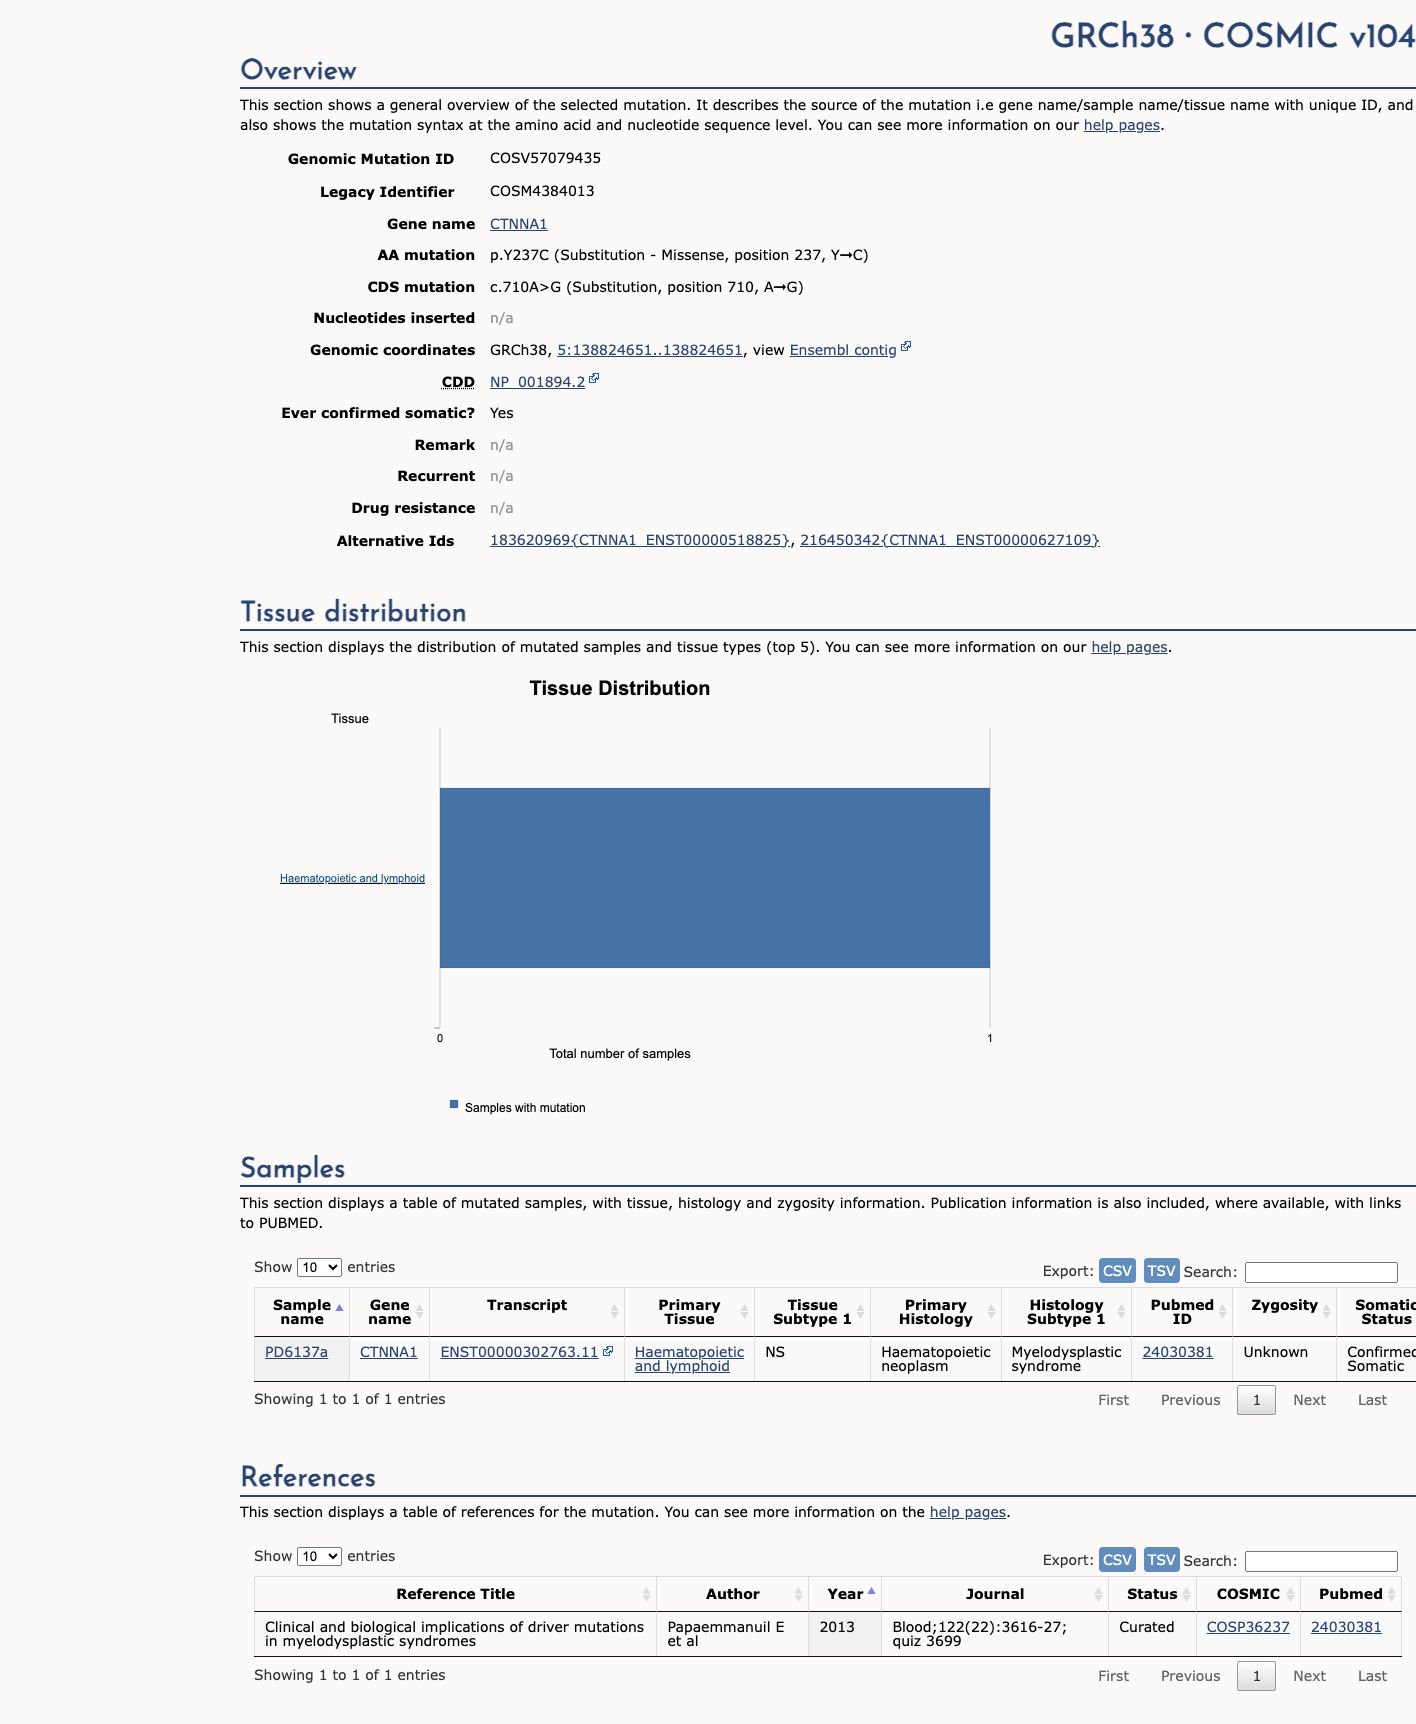
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has previously been reported in somatic cancers (COSMIC; COSV57079435, n = 1 times).
Hotspots
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
1papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 4 further PMIDs triaged but not cited — see Sources & references.
Rule & framework references · cited for criterion definitions, not variant evidence
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 4 PMIDs not cited in assessment
34326862 ↗ Analysis of Sequence and Copy Number Variants in Canadian Patient Cohort With Familial Cancer Syndromes Using a Unique Next Generation Sequencing Based Approach. CLINVAR
25394175 ↗ A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment. CLINVAR
34043773 ↗ European guidelines from the EHTG and ESCP for Lynch syndrome: an updated third edition of the Mallorca guidelines based on gene and gender. CLINVAR
35957908 ↗ Hereditary Breast Cancer in the Brazilian State of Cear&#xe1; (The CHANCE Cohort): Higher-Than-Expected Prevalence of Recurrent Germline Pathogenic Variants. CLINVAR