Back
NM_002691.4:c.3205G>A
p.Val1069Ile · POLD1
ACMG/AMP
0%
complete
Final classification
VUS
PM2BP4
POLD1
c.3205G>A
p.Val1069Ile
missense · exon 26

POLD1 encodes the catalytic subunit of DNA polymerase delta, an enzyme that carries both DNA synthesis and proofreading (3' to 5' exonuclease) activities and is essential for accurate DNA replication and repair. Germline mutations in its exonuclease domain cause polyposis and predispose people to colorectal, endometrial, and possibly brain cancers. In cancer, POLD1 defects impair replication fidelity, leading to the accumulation of many mutations (an ultra-mutated phenotype) that may make tumors more responsive to immunotherapy, though somatic POLD1 mutations are rare.

This variant

POLD1 encodes the catalytic subunit of DNA polymerase delta, whose proofreading activity is required for replication fidelity, and germline POLD1 variants in the exonuclease domain cause dominantly inherited polymerase proofreading-associated polyposis with colorectal and endometrial cancer predisposition; NM_002691.4:c.3205G>A (p.Val1069Ile) alters a residue in the C-terminal CysB region rather than that exonuclease domain, so its relevance to this gene's established cancer-predisposition mechanism remains unproven.
Transcript
NM_002691.4
HGVS · transcript:coding
NM_002691.4:c.3205G>A
GRCh38
chr19:50417256 G>A
GRCh37
chr19:50920513 G>A
VUS: PM2 supporting (very rare in gnomAD) plus BP4 supporting (REVEL 0.197, tolerated) satisfy no ACMG 2015 pathogenic or benign combination threshold.
Classification rationale
PM2 BP4 VUS
POLD1 c.3205G>A missense · exon 26

VUS framework basis: no ClinGen VCEP/CSPEC or local POLD1 framework exists, so the generic ACMG/AMP 2015 combination rules were applied to the adjudicated criteria. PM2 supporting met: gnomAD v4.1 AF 3.76e-06 with popmax 3.02e-05 is far below the 1e-4 rare-variant cutoff. BP4 supporting met: missense variant with REVEL 0.197, at or below the <=0.29 tolerated threshold. PS1 not met: the identical p.Val1069Ile change is only Uncertain significance (2 labs) or Likely benign (1 lab) in ClinVar, never pathogenic. PS3 and BS3 not assessed: no functional assay of POLD1 p.Val1069Ile exists in the indexed literature or any curated database. PS4 not met: no case-control, cohort or penetrance enrichment data for this variant exists. PM1 not met: codon 1069 is not a hotspot and sits in the C-terminal CysB region, outside the cancer-critical exonuclease domain. PM5 not met: the only alternate change at residue 1069, p.Val1069Asp, is Uncertain significance in ClinVar. PP3 not met: REVEL 0.197 is below the >=0.644 supporting threshold for a damaging effect. PP5 and BP6 not met: ClinVar VCV000537093 has zero expert-panel submissions, only conflicting single-submitter records. BA1, BS1 and BS2 not met: frequency is far below 0.05/0.01 thresholds, with no homozygotes or phenotyped healthy carriers. PS2, PM6, PP1, BS4 and PP4 not assessed; PM3, PM4, PVS1, BP3, BP7 not applicable; PP2, BP1, BP2 and BP5 not met - none contribute evidence.

PM2 + BP4 VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_002691.4 · variants mapped to exon structure
POLD1 NM_002691.4
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 21 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
PM2 supporting Pathogenic
Met at supporting strength: gnomAD v4.1 frequency 3.76e-06 with grpmax FAF 1.25e-06 and popmax 3.02e-05 falls below the 1e-4 PM2 threshold.
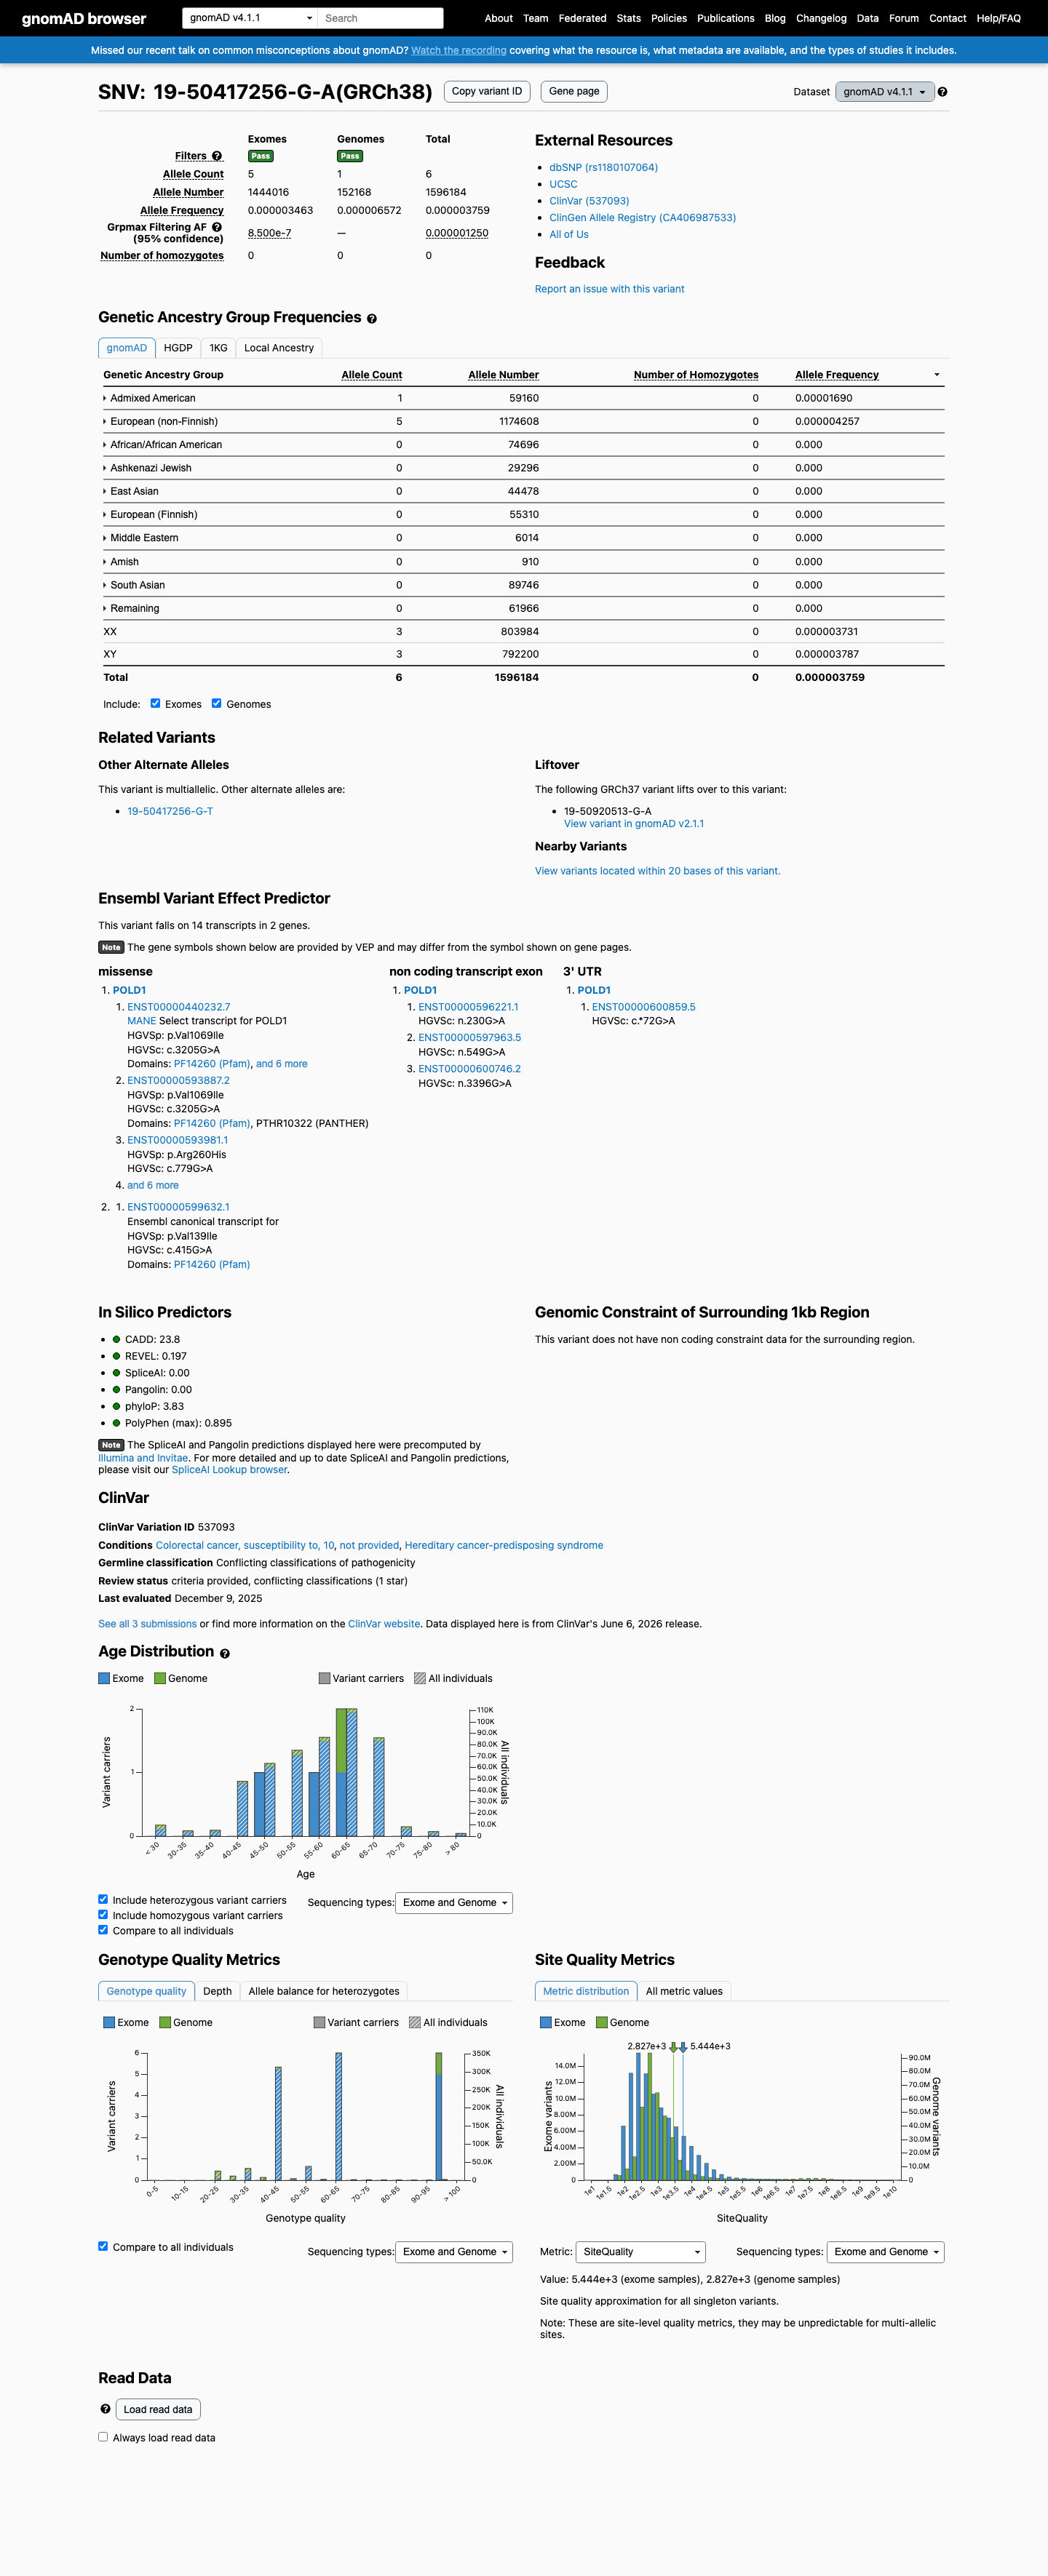
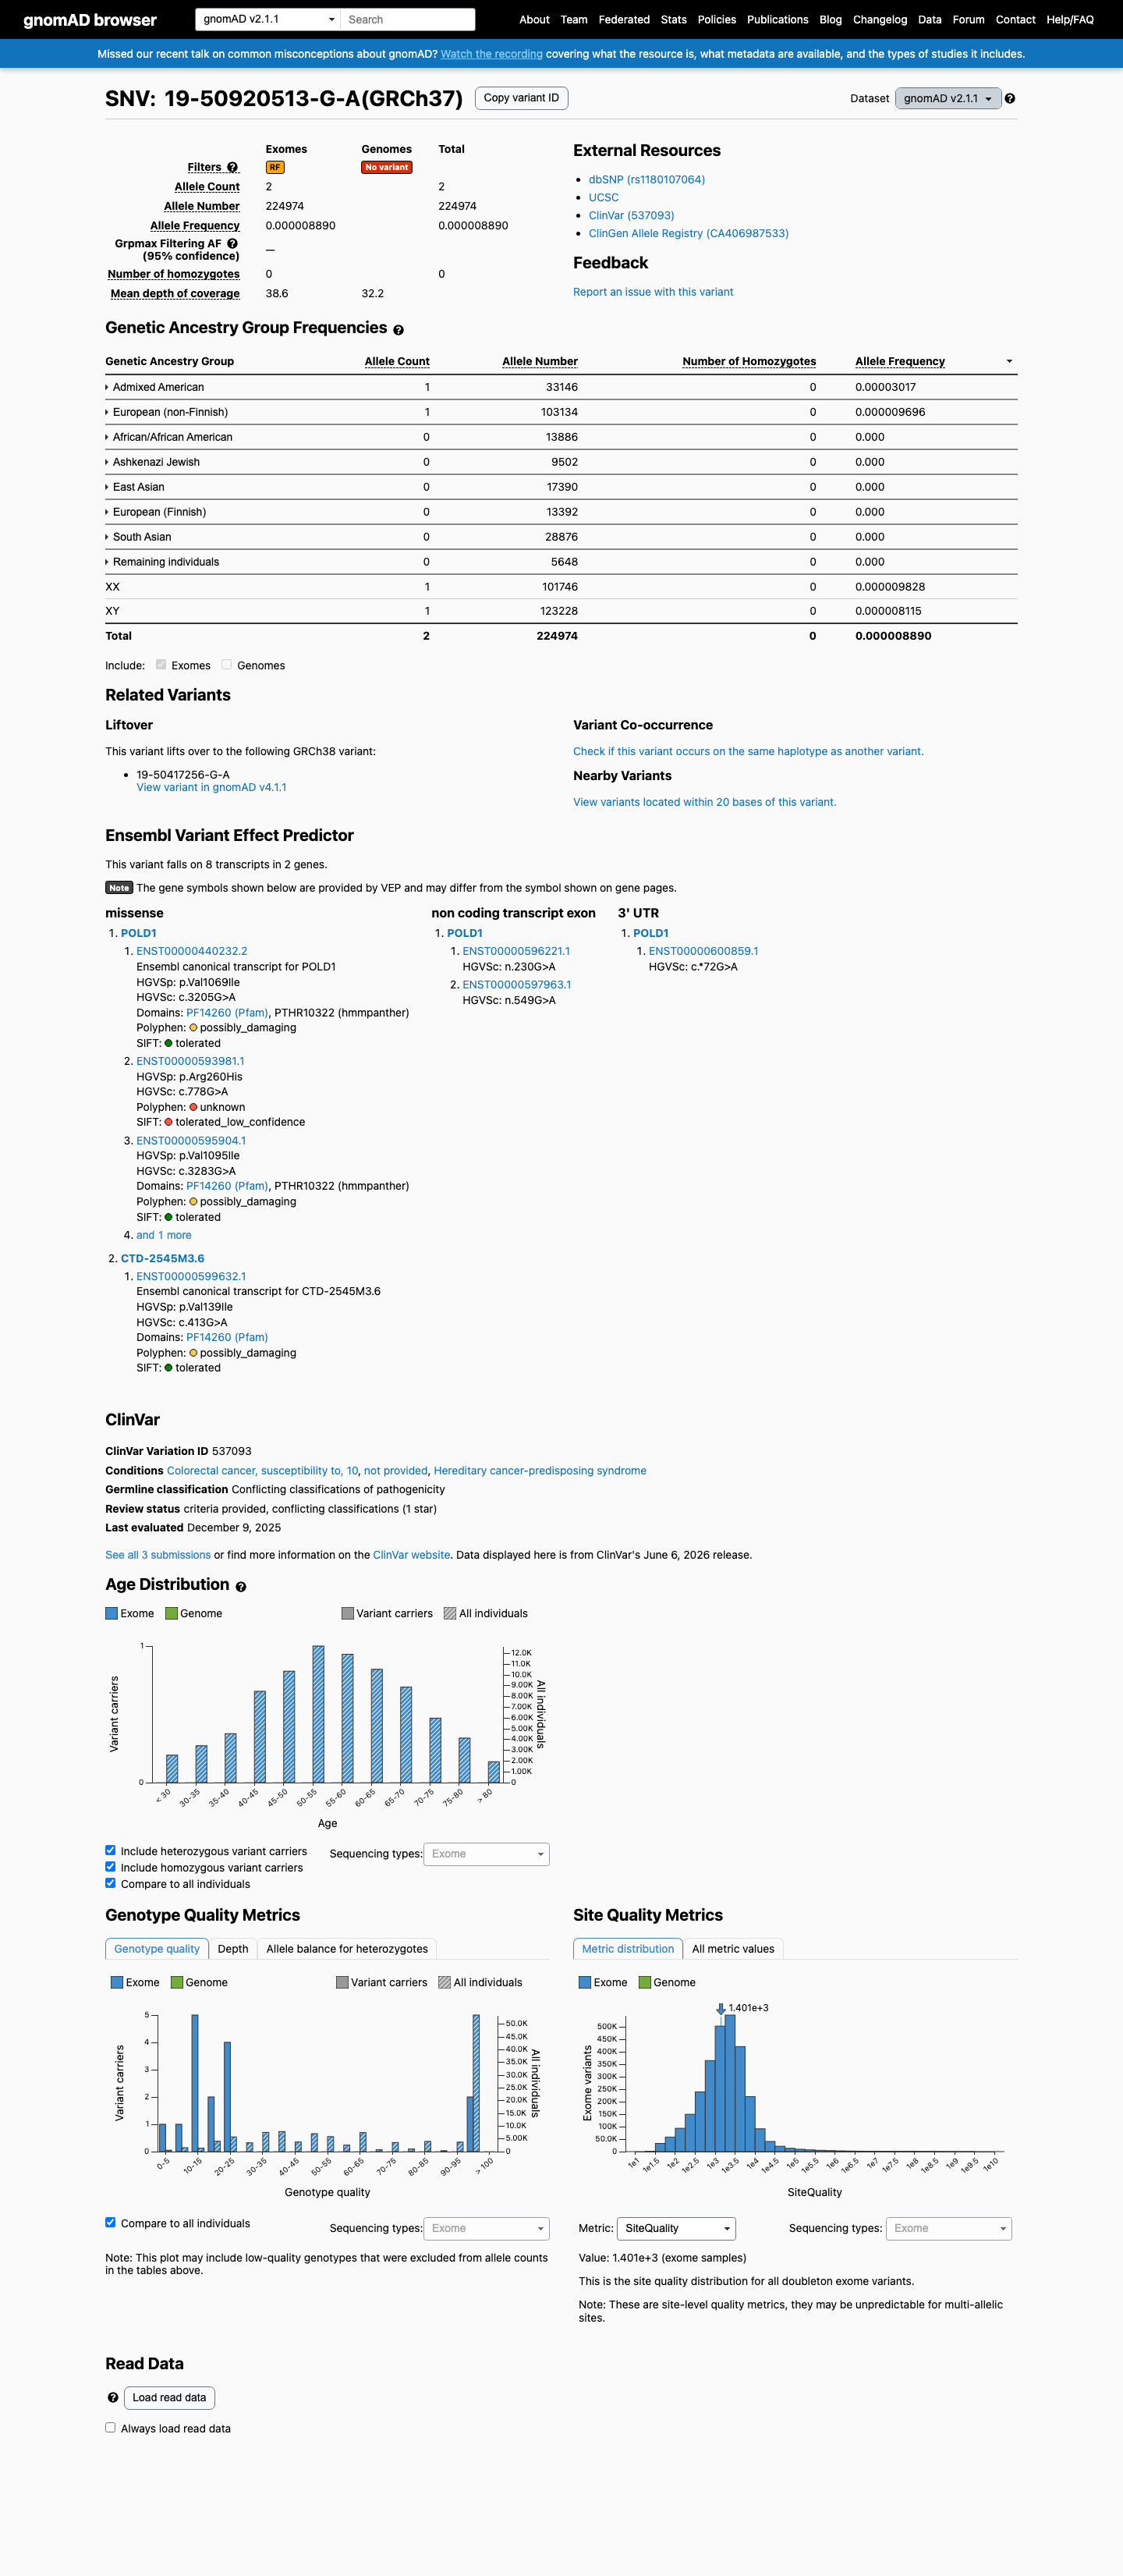
gnomAD v4.1 joint (exomes + genomes, all-comers default source): AF 3.75897e-06 (6/1,596,184 alleles), 0 homozygotes, grpmax filtering allele frequency 1.25e-06; exome component 3.46257e-06 (5/1,444,016) and genome component 6.57168e-06 (1/152,168).gnomAD v2.1 (exomes only, all-comers default source): AF 8.88992e-06 (2/224,974 alleles), 0 homozygotes; no grpmax filtering allele frequency reported for this variant.Highest observed ancestry-group frequency: Admixed American, gnomAD v2.1 AF 3.01696e-05 (1/33,146 alleles, 0 homozygotes) and gnomAD v4.1 AF 1.69033e-05 (1/59,160 alleles, 0 homozygotes).
BP4 supporting Benign
Met at supporting strength: REVEL 0.197 is at or below the <=0.29 BP4 threshold.
Variant consequence: NM_002691.4:c.3205G>A encodes NP_002682.2:p.(Val1069Ile), a missense substitution; BP4 therefore draws on the REVEL missense path and not on SpliceAI.No POLD1 ClinGen VCEP specification or gene-specific PP3/BP4 lookup table was available, so generic ACMG/AMP computational rules were applied.REVEL v1.3 local predictor lookup (source_registry key 'revel') returned score 0.197; ClinGen SVI REVEL calibration places BP4 supporting at <=0.29, moderate at <=0.183, strong at <=0.016 (Pejaver et al. 2022, Am J Hum Genet, PMID:36413997). 0.197 satisfies supporting but not moderate or strong.
Assessed · not applied · 13 not met · 8 not assessed
Pathogenic
PS1 Not met: the identical p.Val1069Ile change is classified only Uncertain significance (2 labs) or Likely benign (1 lab) in ClinVar, never pathogenic.
PS2 Not assessed: this record contains no proband, trio, or parental-testing data, so a confirmed de novo occurrence of c.3205G>A cannot be established.
PS3 Not assessed: no functional study of POLD1 p.Val1069Ile exists - PubMed/PMC searches for V1069I, Val1069Ile and c.3205G>A returned zero records.
PS4 Not met: no case-control or cohort enrichment data exists for p.Val1069Ile, and the only triaged paper (PMID:25394175) never mentions POLD1.
PM1 Not met: codon 1069 is not a hotspot and sits in the C-terminal CysB region, outside POLD1's established cancer-critical exonuclease domain.
PM5 Not met: the only alternate change at residue 1069, p.Val1069Asp, is classified Uncertain significance in ClinVar, so no pathogenic comparator exists.
PM6 Not assessed: no proband or parental data is present in this record, so an assumed de novo occurrence of c.3205G>A cannot be asserted.
PP1 Not assessed: no pedigree or relative genotypes exist for this case, so co-segregation of c.3205G>A cannot be counted across any meioses.
PP2 Not met: gnomAD v4 shows POLD1 is not missense-constrained (o/e 0.84, missense Z 2.75 versus the 3.09 threshold), so benign missense variation is not rare.
PP3 Not met: missense variant, REVEL 0.197 is below the >=0.644 PP3 supporting threshold.
PP4 Not assessed: no proband phenotype or family history is available, and the ClinVar condition labels are broad cancer-predisposition categories, not a single-gene-specific phenotype.
PP5 Not met: ClinVar VCV000537093 has zero expert-panel submissions, only conflicting single-submitter records (two uncertain significance, one likely benign), so the expert-panel requirement fails.
Benign
BA1 Not met: the highest observed population frequency, 3.02e-05 in gnomAD v2.1 Admixed Americans, is over three orders of magnitude below the 0.05 BA1 threshold.
BS1 Not met: the highest observed allele frequency, 3.02e-05, sits about 300-fold below the 0.01 BS1 threshold and no gene-specific threshold was derivable.
BS2 Not met: zero homozygotes among 1,596,184 gnomAD v4.1 alleles and no phenotyped healthy adult carriers, and the gene's adult-onset incomplete penetrance defeats the BS2 premise.
BS3 Not assessed: zero PubMed/PMC records exist for POLD1 p.Val1069Ile (V1069I, Val1069Ile, c.3205G>A), so no assay can show a non-damaging effect.
BS4 Not assessed: no family genotype data exists in this record, so non-segregation of c.3205G>A with disease cannot be demonstrated.
BP1 Not met: POLD1 disease is caused by missense variants (exonuclease-domain polyposis; CysB-region MDPL and immunodeficiency), not primarily by truncating variants.
BP2 Not met: no pathogenic POLD1 variant was observed in cis or trans with p.Val1069Ile in any ClinVar submission or literature source examined.
BP5 Not assessed: no proband-level genetic data or case report exists to show, or exclude, an alternate molecular basis for disease in a carrier of p.Val1069Ile.
BP6 Not met: the only benign-direction ClinVar record is a single non-expert Ambry Genetics Likely benign submission (expert_panel_submissions = 0), which cannot trigger BP6.
N/A · 5 PVS1 · PM3 · PM4 · BP3 · BP7
Research & evidence
Population frequency
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 3.75897e-06; MAF= 0.00038%, 6/1596184 alleles, homozygotes = 0) and has highest observed frequency in the Admixed American population (AF= 1.69033e-05; MAF= 0.00169%, 1/59160 alleles, homozygotes = 0); grpmax FAF= 1.25e-06.
v2.1
This variant is present in gnomAD v2.1 (AF= 8.88992e-06; MAF= 0.00089%, 2/224974 alleles, homozygotes = 0) and has highest observed frequency in the Admixed American population (AF= 3.01696e-05; MAF= 0.00302%, 1/33146 alleles, homozygotes = 0).
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.00038% · 6 / 1,596,184
0 hom · FAF 0.00013%
Admixed American
1 / 59,160
0.0017%
European (non-Finnish)
5 / 1,174,608
0.00043%
+ 8 not observed (Remaining individuals, European (Finnish), Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American)
gnomAD v2.1
0.00089% · 2 / 224,974
0 hom
Admixed American
1 / 33,146
0.003%
European (non-Finnish)
1 / 103,134
0.00097%
+ 6 not observed (African/African American, Ashkenazi Jewish, East Asian, European (Finnish), Remaining individuals, South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
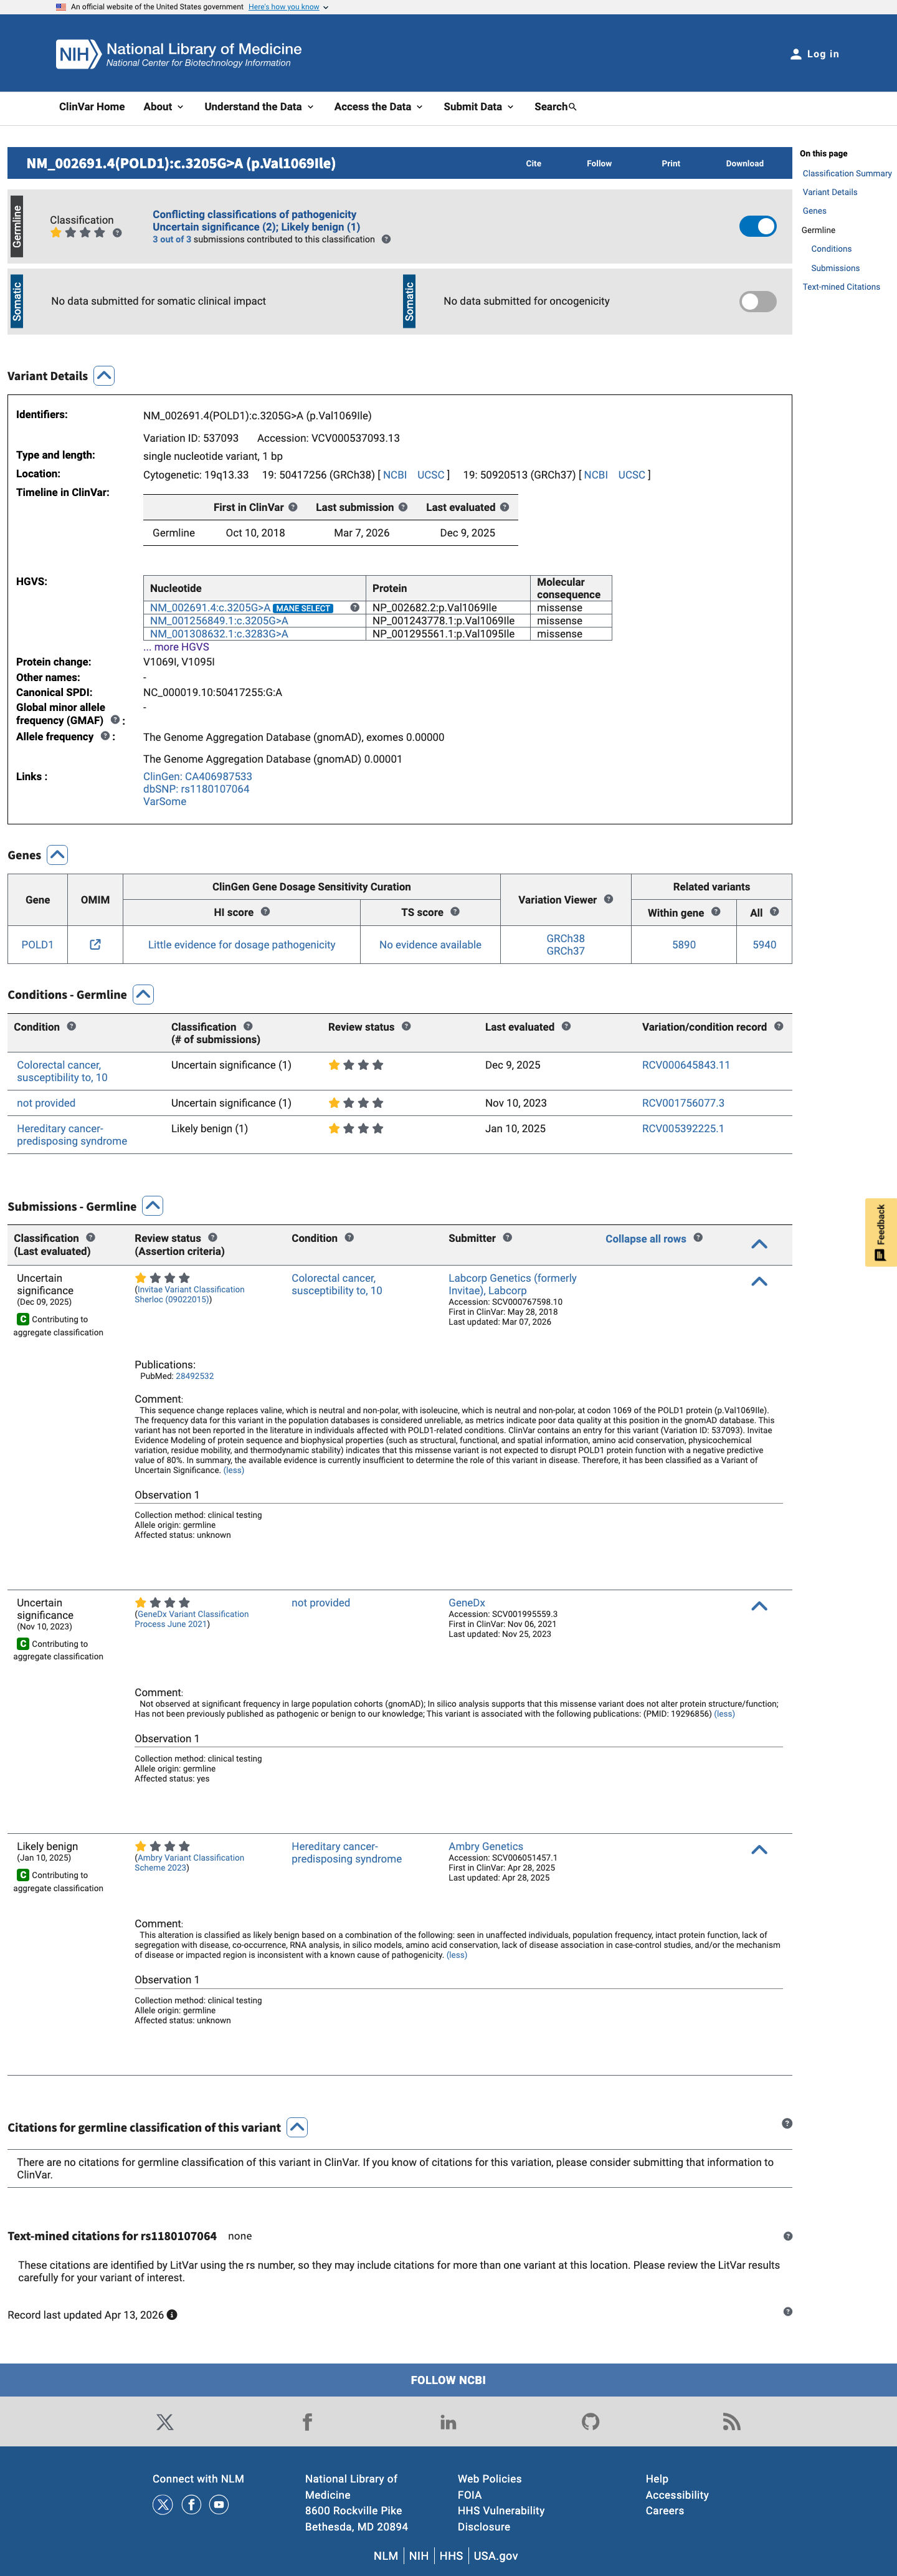
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Uncertain significance (2 clinical laboratories) and as Likely benign (1 clinical laboratory). (ClinVarID = 537093)
SpliceAI screenshot
In silico
SpliceAI returned NO scores for this variant, so no SpliceAI-based splice prediction is available. This is missing data, NOT evidence of absent splice impact: it must not be used to support BP4 or to argue against PP3/PVS1. Pangolin scores may be present but are not calibrated for PP3/BP4 here. REVEL score = 0.197. BayesDel score = -0.367908.
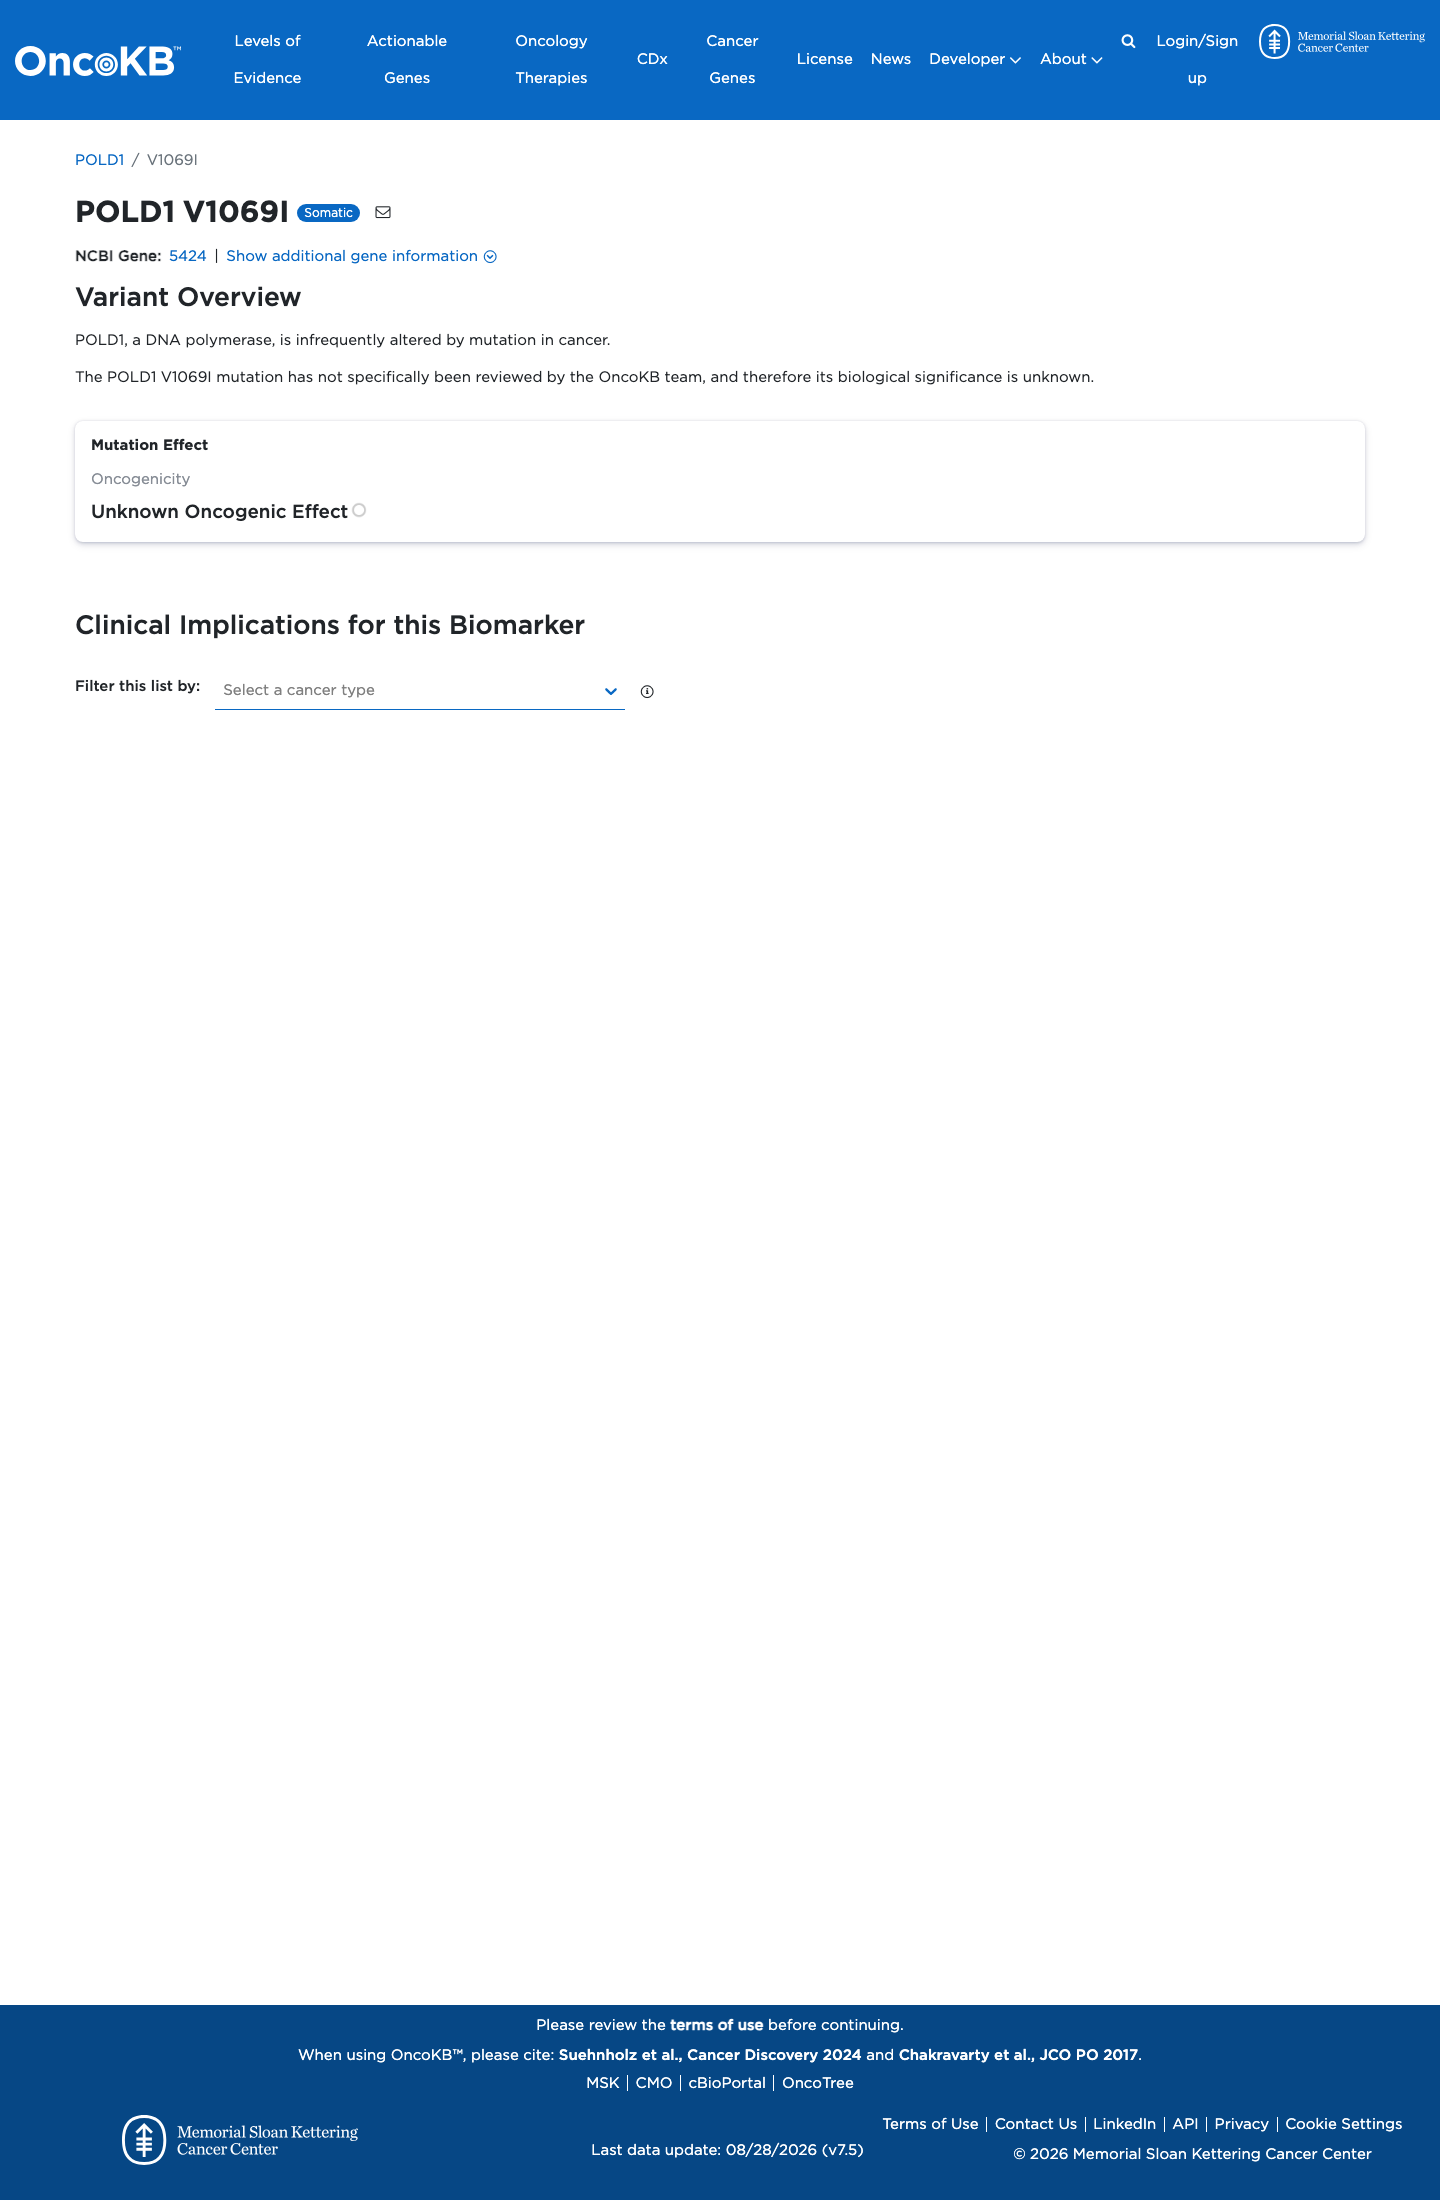
Functional / OncoKB screenshot
Functional Unknown Oncogenic Effect
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. POLD1, a DNA polymerase, is infrequently altered by mutation in cancer.
OncoKB ↗
COSMIC screenshot
COSMIC
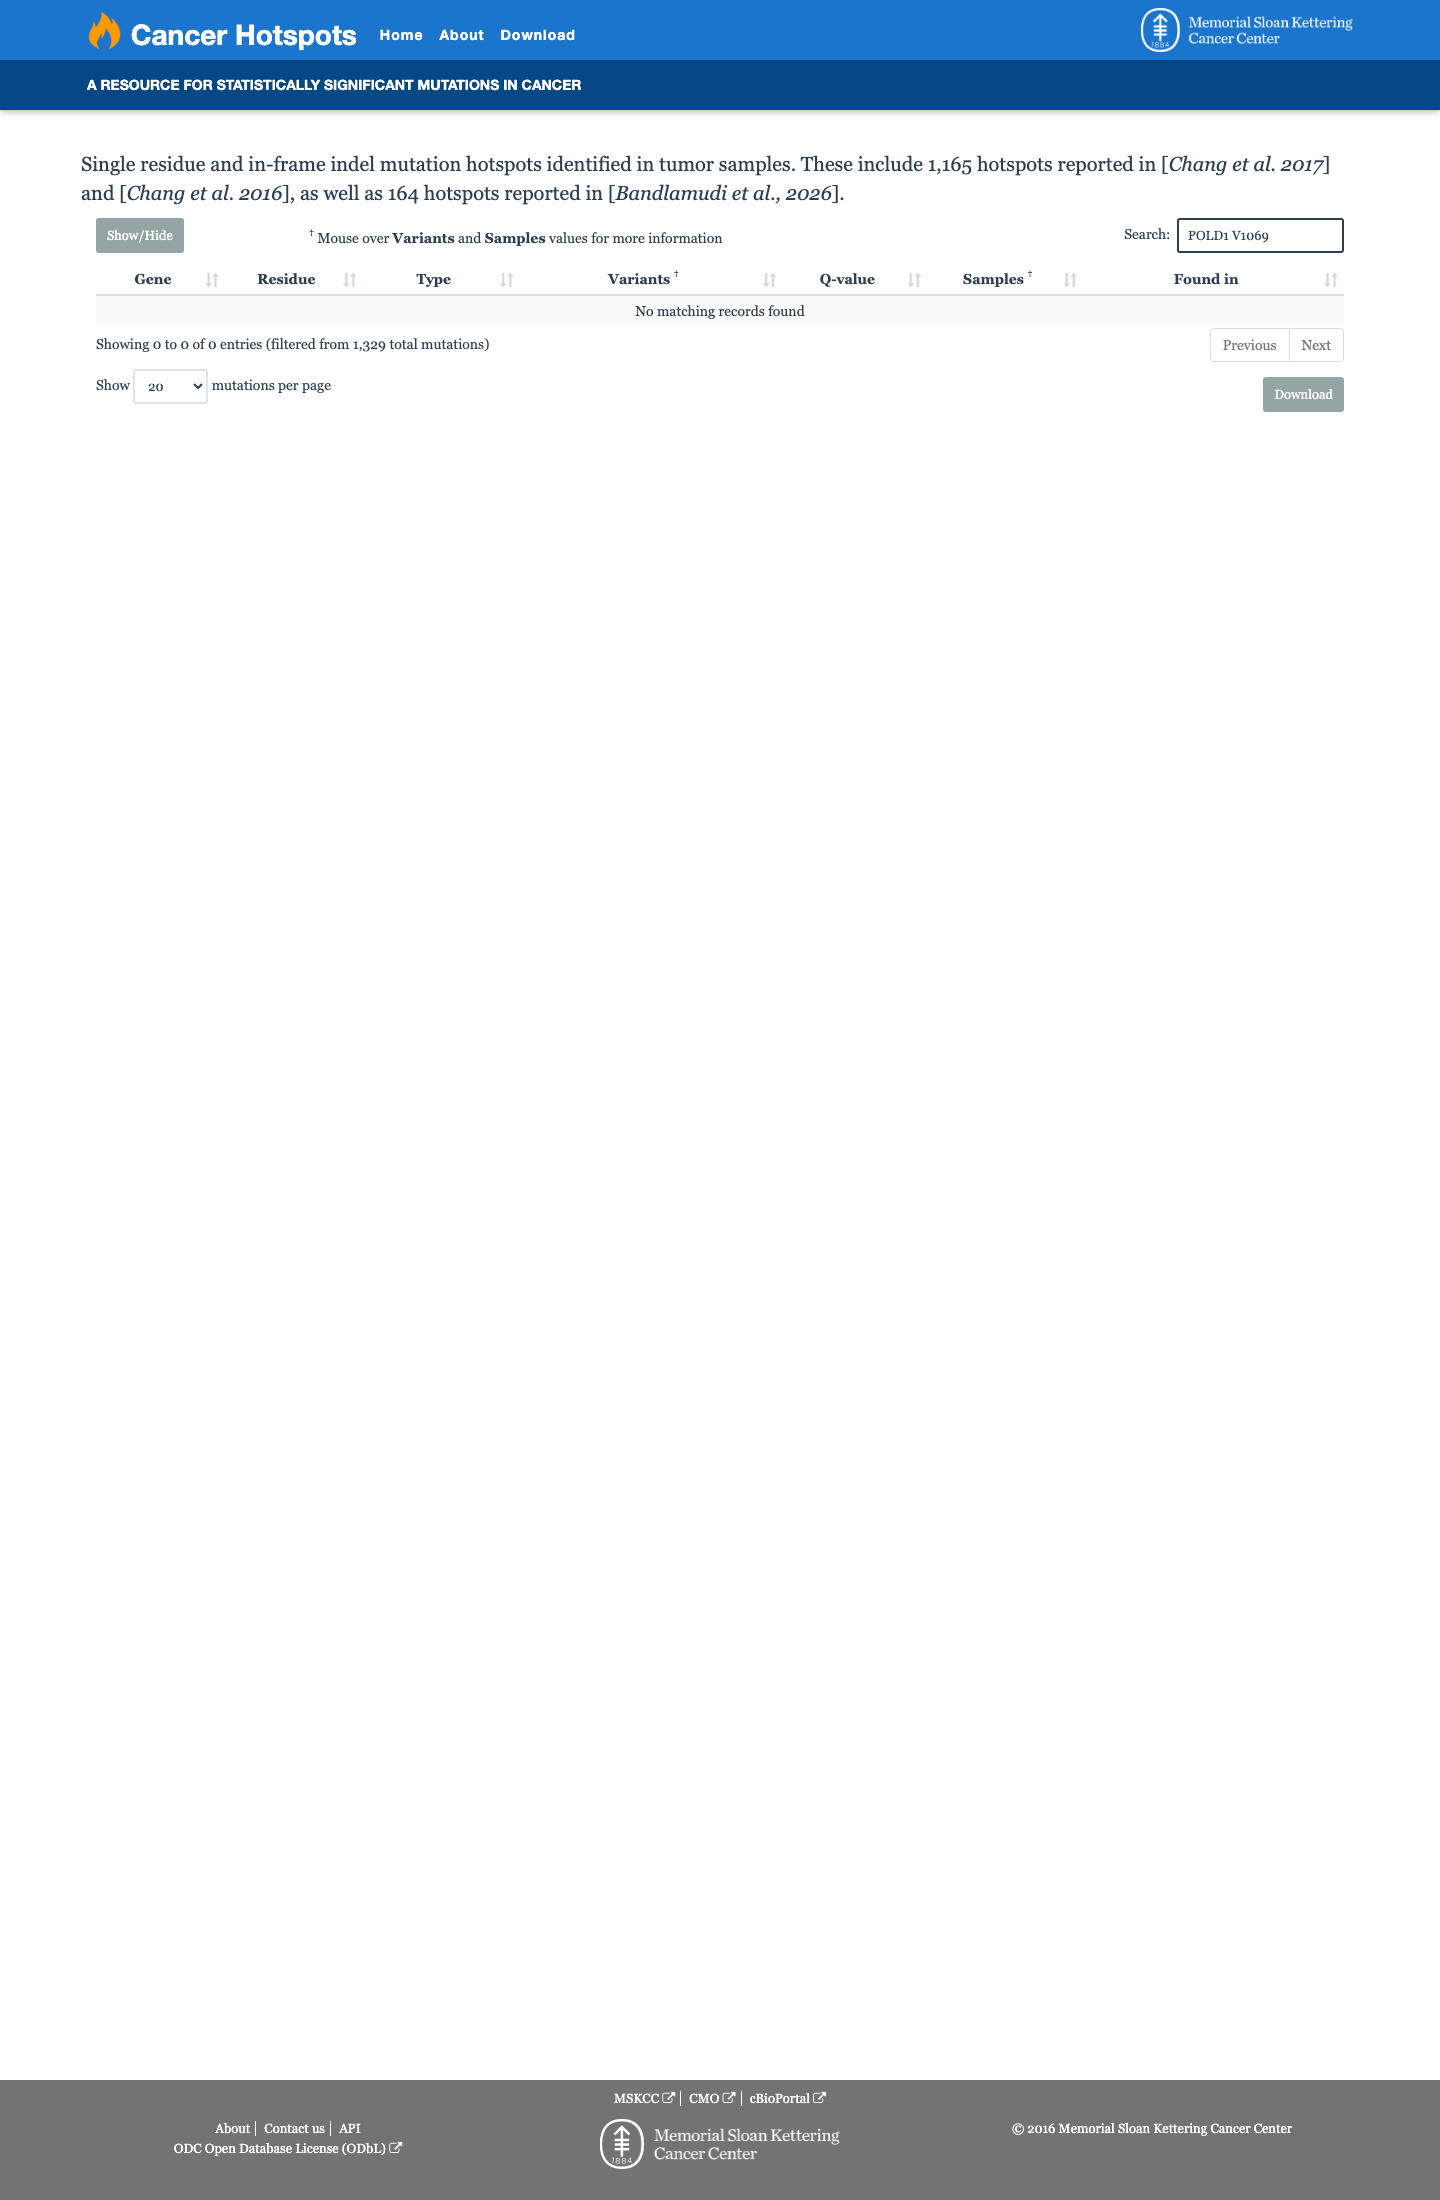
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
1papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why.
Rule & framework references · cited for criterion definitions, not variant evidence
25394175 ↗ A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots