Back
NM_001042492.2:c.4375_4377del
p.Glu1459del · NF1
0%
complete
Final classification
VUS
PM2PM4
NF1
c.4375_4377del
p.Glu1459del
in_frame_indel_unknown · exon 33

NF1 encodes neurofibromin, a tumor suppressor protein that negatively regulates the RAS signal transduction pathway. As a GTPase-activating protein, it helps switch RAS proteins from their active to inactive state, keeping cell growth in check; loss of NF1 function leaves RAS overactive and drives downstream growth pathways such as MAPK/ERK and PI3K. Inherited changes in NF1 cause the cancer-predisposition syndrome neurofibromatosis type 1, and are also linked to juvenile myelomonocytic leukemia and Watson syndrome. Somatic changes in NF1 are found in many tumor types, including breast cancer, melanoma, and glioma.

This variant

This NF1 in-frame deletion changes neurofibromin, a tumor suppressor that normally restrains RAS signaling and thereby helps regulate cell growth.

Transcript
NM_001042492.2
HGVS · transcript:coding
NM_001042492.2:c.4375_4377del
GRCh38
chr17:31259068 AAAG>A
GRCh37
chr17:29586086 AAAG>A
VUS: PM4 (moderate) plus PM2 (supporting) do not satisfy a generic ACMG/AMP Likely Pathogenic or Pathogenic combination.
Classification rationale
PM2PM4 VUS
NF1 c.4375_4377del in_frame_indel_unknown · exon 33

PM2 supporting: the variant is absent from gnomAD v2.1 and v4.1. PM4 moderate: the 3-bp in-frame deletion produces p.(Glu1459del), a one-amino-acid protein-length change.

PM2 + PM4 → VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_001042492.2 · variants mapped to exon structure
NF1 NM_001042492.2
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 16 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
PM2 supporting Pathogenic
Met at supporting strength: the variant is absent from gnomAD v2.1 and v4.1, giving frequency 0 versus the PM2 threshold of <=0.0001.
The NF1 ClinGen specification was reviewed first, but it provides no usable PM2 population-source restriction or threshold.gnomAD v2.1 reports the variant as absent, including the available non-cancer check.gnomAD v4.1 reports the variant as absent, and the available gnomAD v3.1 non-cancer check also reports absence.
PM4 moderate Pathogenic
Met: NM_001042492.2:c.4375_4377del is a 3-bp in-frame deletion producing p.(Glu1459del), a one-amino-acid protein-length change meeting generic PM4.
Sequence normalization maps NM_001042492.2:c.4375_4377del to NP_001035957.1:p.(Glu1459del), a 3-bp in-frame deletion removing one amino acid.Variant Validator places the deletion in NF1 exon 33 and confirms the predicted protein consequence p.(Glu1459del).The retrieved NF1 ClinGen framework is incomplete and provides no structured PM4 rule, so the generic ACMG/AMP protein-length-change rule is used.
Assessed · not applied · 6 not met · 10 not assessed
Pathogenic
PS2 Not assessed: ClinVar reports one de novo case with confirmed parentage, but the candidate record is not an exact match and parental genotypes are unavailable.
PS3 Not assessed: no variant-specific validated functional assay or measured effect was reported for p.(Glu1459del).
PS4 Not met: no case-control enrichment, odds ratio, likelihood ratio, or statistically significant phenotype prevalence was reported for exact variant c.4375_4377del.
PM6 Not assessed: an apparently de novo claim lacks verifiable exact-variant documentation and the parental-testing details required for PM6.
PP1 Not assessed: no variant-positive affected relatives or informative cosegregating meioses are documented for this exact NF1 deletion.
PP4 Not assessed: no patient-specific, highly specific NF1 phenotype was documented to support PP4 for this exact variant.
PP5 Not met: ClinVar has no exact-variant expert-panel pathogenic or likely pathogenic assertion; available entries are non-expert submissions or aggregate labels.
Benign
BA1 Not met: the variant is absent from gnomAD v2.1 and v4.1, giving observed frequency 0 versus the BA1 threshold of >=0.05.
BS1 Not met: the variant is absent from gnomAD, with observed frequency 0 versus the generic BS1 threshold of >=0.01.
BS2 Not assessed: gnomAD absence provides no qualifying observation of the variant in healthy individuals or homozygotes for BS2.
BS3 Not assessed: no variant-specific validated assay demonstrated normal NF1 function for p.(Glu1459del).
BS4 Not assessed: no unaffected variant carriers or other informative non-segregation observations are documented for this exact deletion.
BP2 Not assessed: no affected-proband data or cis/trans phase result with another pathogenic NF1 variant is available for this deletion.
BP3 Not met: the one-amino-acid in-frame deletion is described as occurring in a non-repeat region, so BP3's repeat-region requirement is not satisfied.
BP5 Not assessed: no case-specific alternative molecular etiology was documented, and no NF1-specific BP5 threshold was available for comparison.
BP6 Not met: no exact-variant ClinVar expert-panel benign or likely benign classification was found; available submissions are non-expert pathogenic or likely pathogenic assertions.
N/A · 10 PVS1 · PS1 · PM1 · PM3 · PM5 · PP2 · PP3 · BP1 · BP4 · BP7
Research & evidence
Population frequency
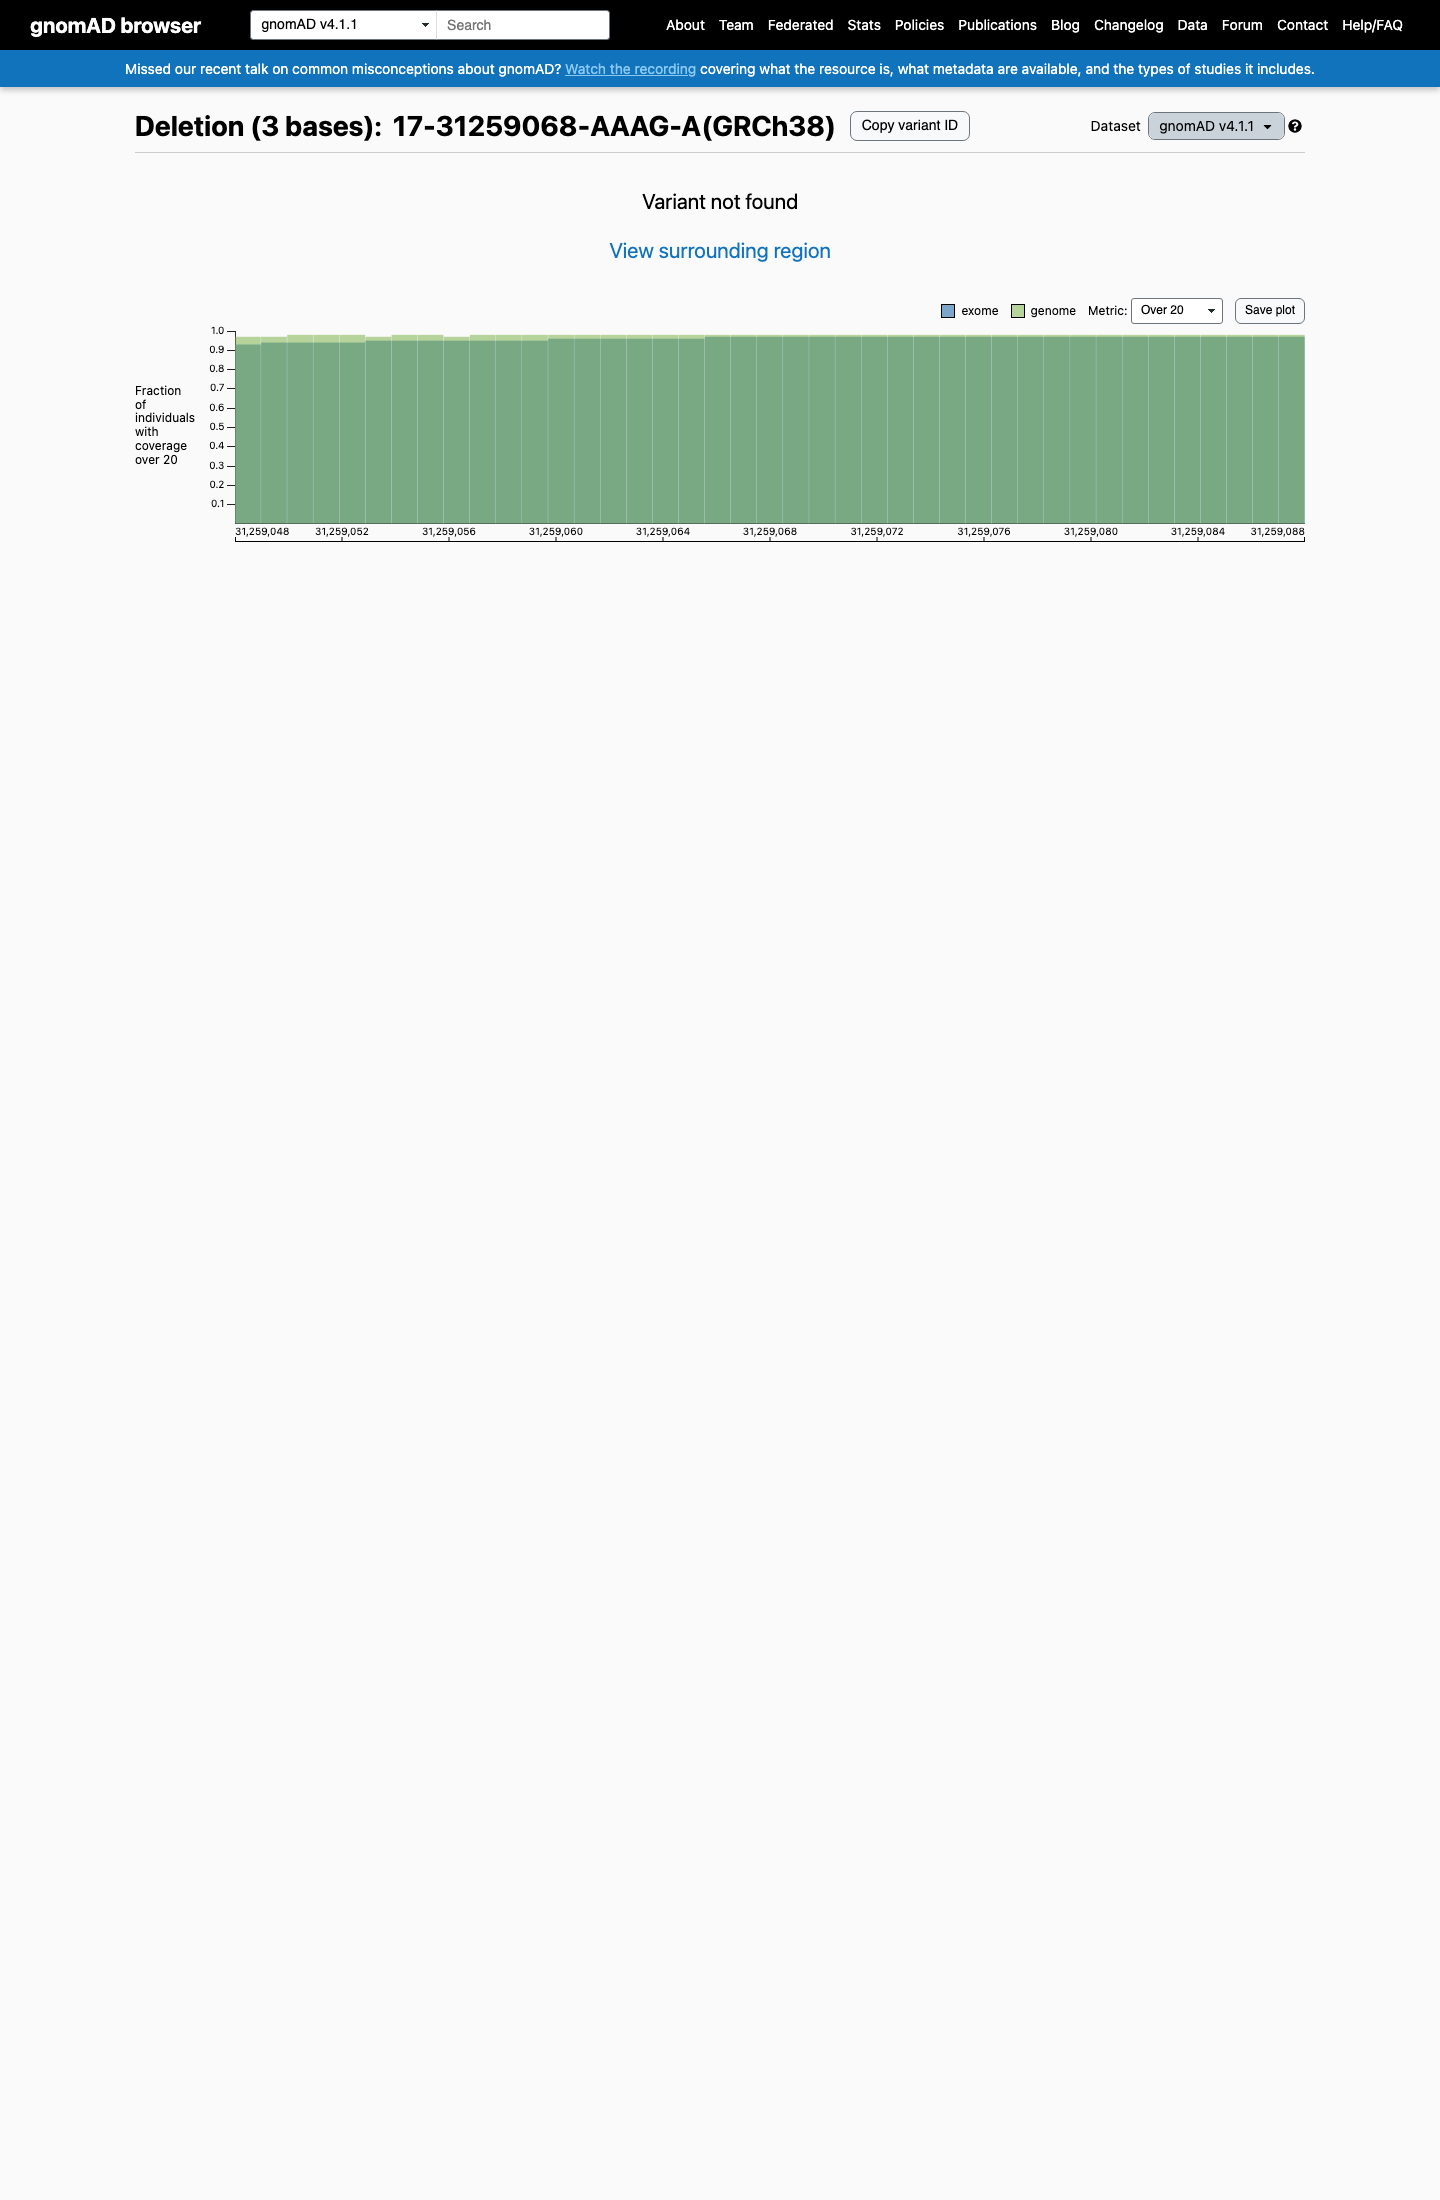
gnomAD v4.1 screenshot
gnomAD v4.1
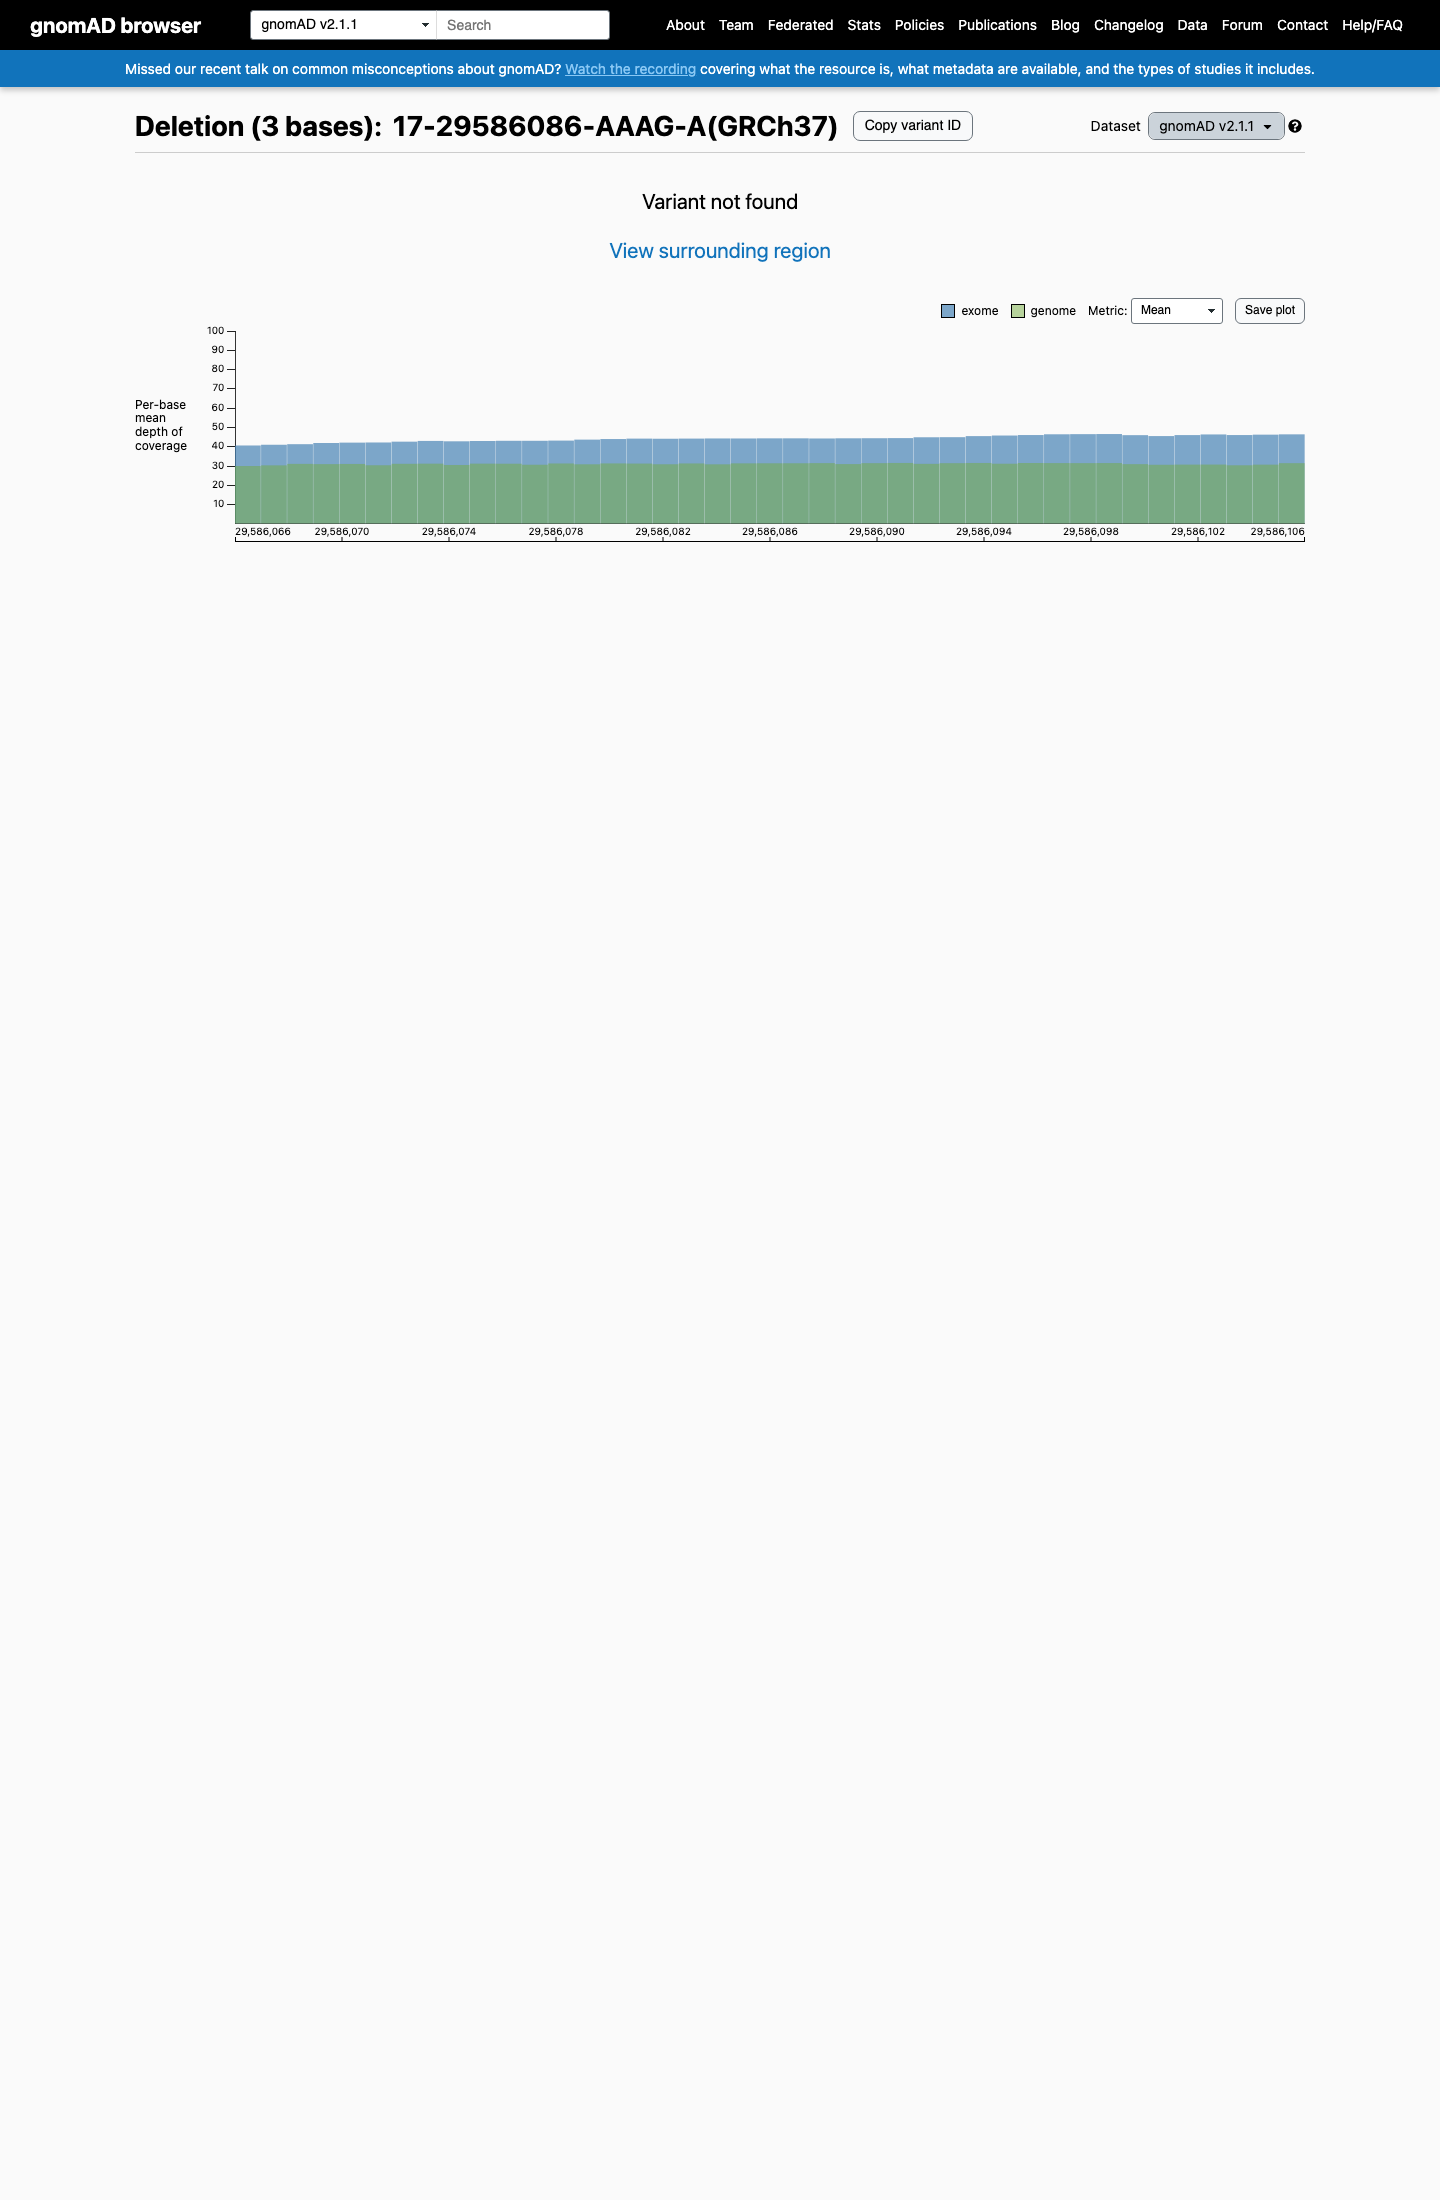
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
Absent from gnomAD v4.1.
v2.1
Absent from gnomAD v2.1.
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
gnomAD v2.1
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
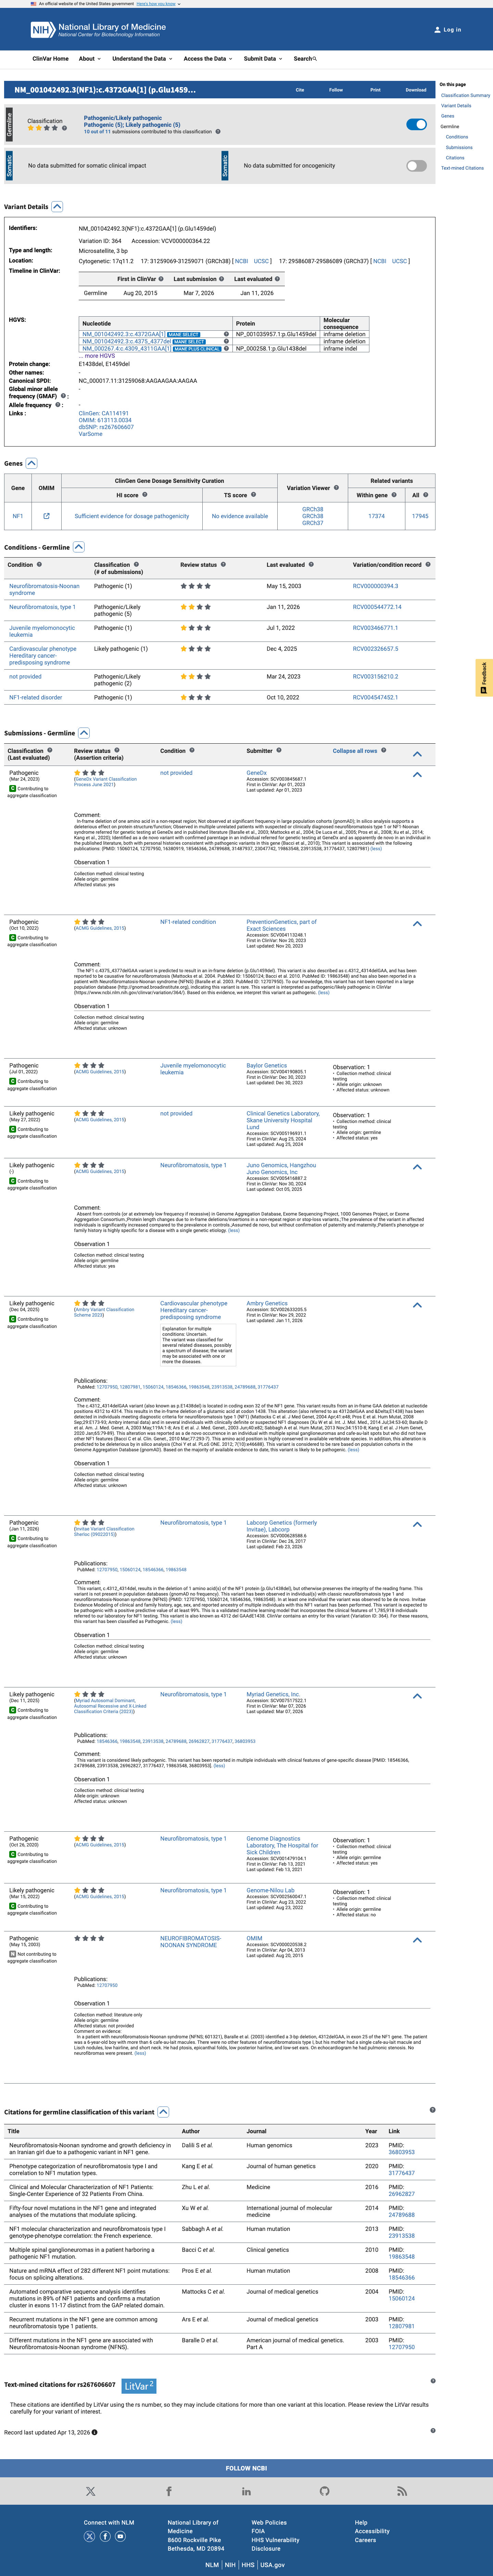
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Pathogenic (5 clinical laboratories) and as Likely pathogenic (5 clinical laboratories). (ClinVarID = 364)
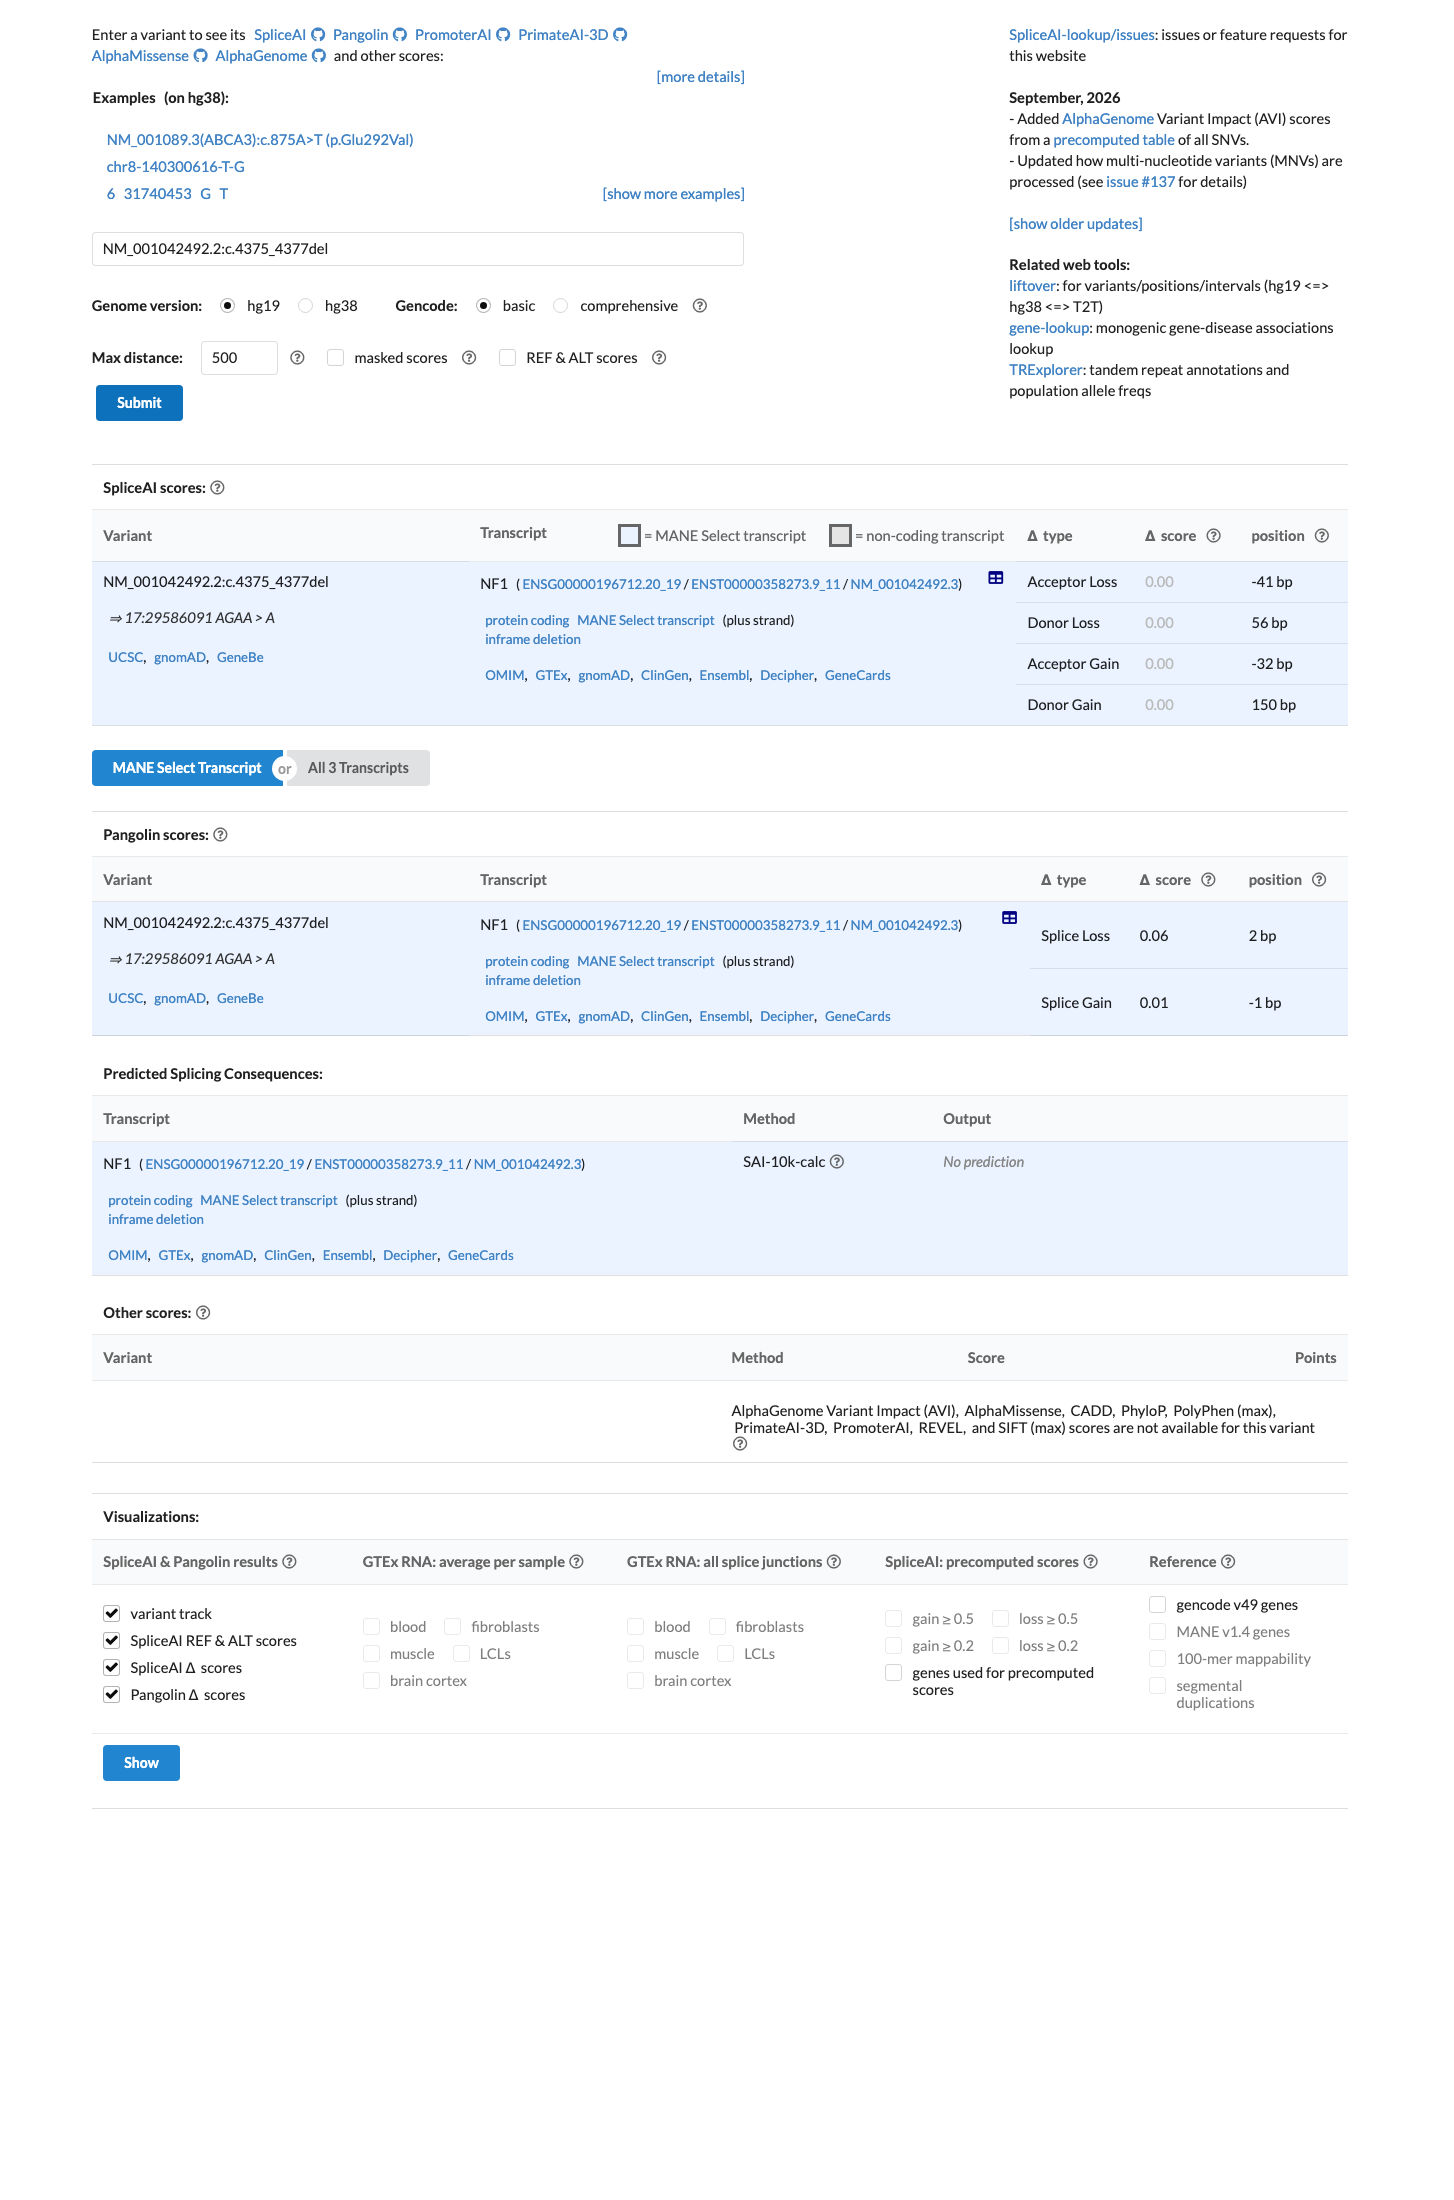
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.00).
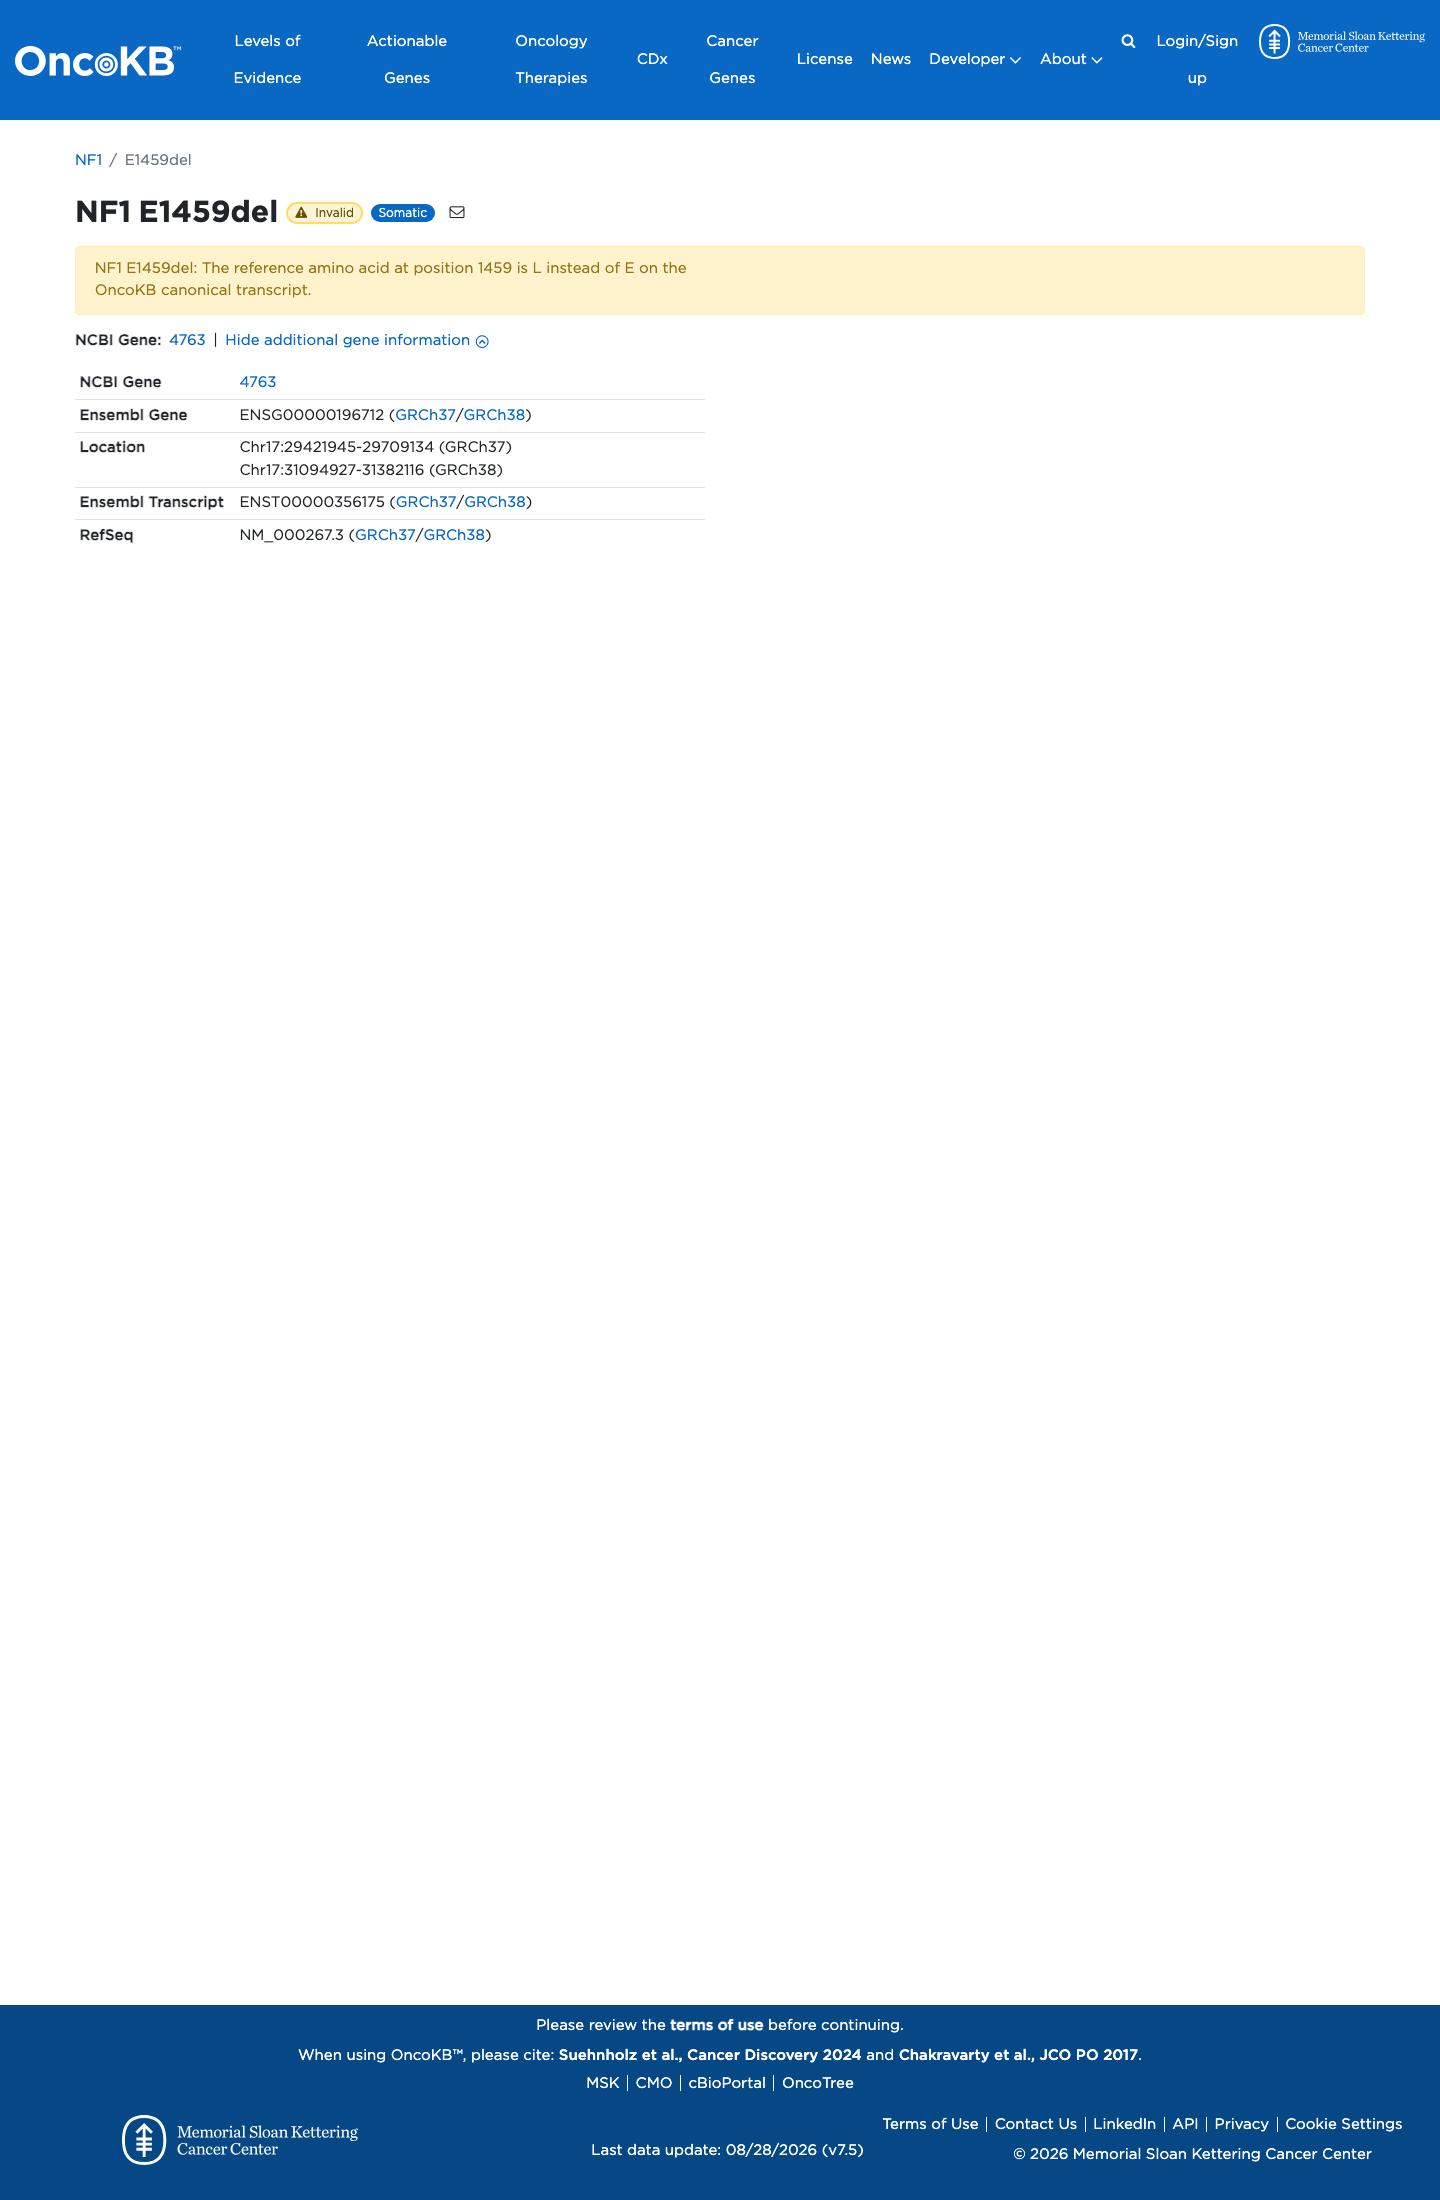
Functional / OncoKB screenshot
Functional
OncoKB identified curated literature and non-variant-specific oncogenicity context for review; listed oncogenicity label: not classified.
OncoKB ↗
COSMIC screenshot
COSMIC
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Sources & reference links
9Sources
CSpec VCEP
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 10 PMIDs not cited in assessment
26962827 ↗ Clinical and Molecular Characterization of NF1 Patients: Single-Center Experience of 32 Patients From China. CLINVAR
31776437 ↗ Phenotype categorization of neurofibromatosis type I and correlation to NF1 mutation types. CLINVAR
12807981 ↗ Recurrent mutations in the NF1 gene are common among neurofibromatosis type 1 patients. CLINVAR
15060124 ↗ Automated comparative sequence analysis identifies mutations in 89% of NF1 patients and confirms a mutation cluster in exons 11-17 distinct from the GAP related domain. CLINVAR
15604628 ↗ Genetic cancer risk assessment and counseling: recommendations of the national society of genetic counselors. CLINVAR
20301471 ↗ Wilms Tumor Predisposition. CLINVAR
20664475 ↗ The North American Neuroendocrine Tumor Society consensus guideline for the diagnosis and management of neuroendocrine tumors: pheochromocytoma, paraganglioma, and medullary thyroid cancer. CLINVAR
26324357 ↗ American Society of Clinical Oncology Policy Statement Update: Genetic and Genomic Testing for Cancer Susceptibility. CLINVAR
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification criteria. CLINVAR
33939658 ↗ The North American Neuroendocrine Tumor Society Consensus Guidelines for Surveillance and Management of Metastatic and/or Unresectable Pheochromocytoma and Paraganglioma. CLINVAR