NF1 encodes neurofibromin, a tumor suppressor protein that negatively regulates the RAS signal transduction pathway. As a GTPase-activating protein, it helps switch RAS proteins from their active to inactive state, keeping cell growth in check; loss of NF1 function leaves RAS overactive and drives downstream growth pathways such as MAPK/ERK and PI3K. Inherited changes in NF1 cause the cancer-predisposition syndrome neurofibromatosis type 1, and are also linked to juvenile myelomonocytic leukemia and Watson syndrome. Somatic changes in NF1 are found in many tumor types, including breast cancer, melanoma, and glioma.
This variant
This NF1 variant occurs in a gene encoding neurofibromin, a RAS-pathway tumor suppressor whose loss of function contributes to neurofibromatosis type 1 and related cancer predisposition.
Transcript
NM_001042492.2
HGVS · transcript:coding
NM_001042492.2:c.1806A>G
GRCh38
chr17:31223528 A>G
GRCh37
chr17:29550546 A>G
VUS: BP7 (supporting) is the only applied criterion; no pathogenic, likely pathogenic, benign, or likely benign ACMG combination is met.
Classification rationale
BP7VUS
NF1 c.1806A>Gsynonymous · exon 16
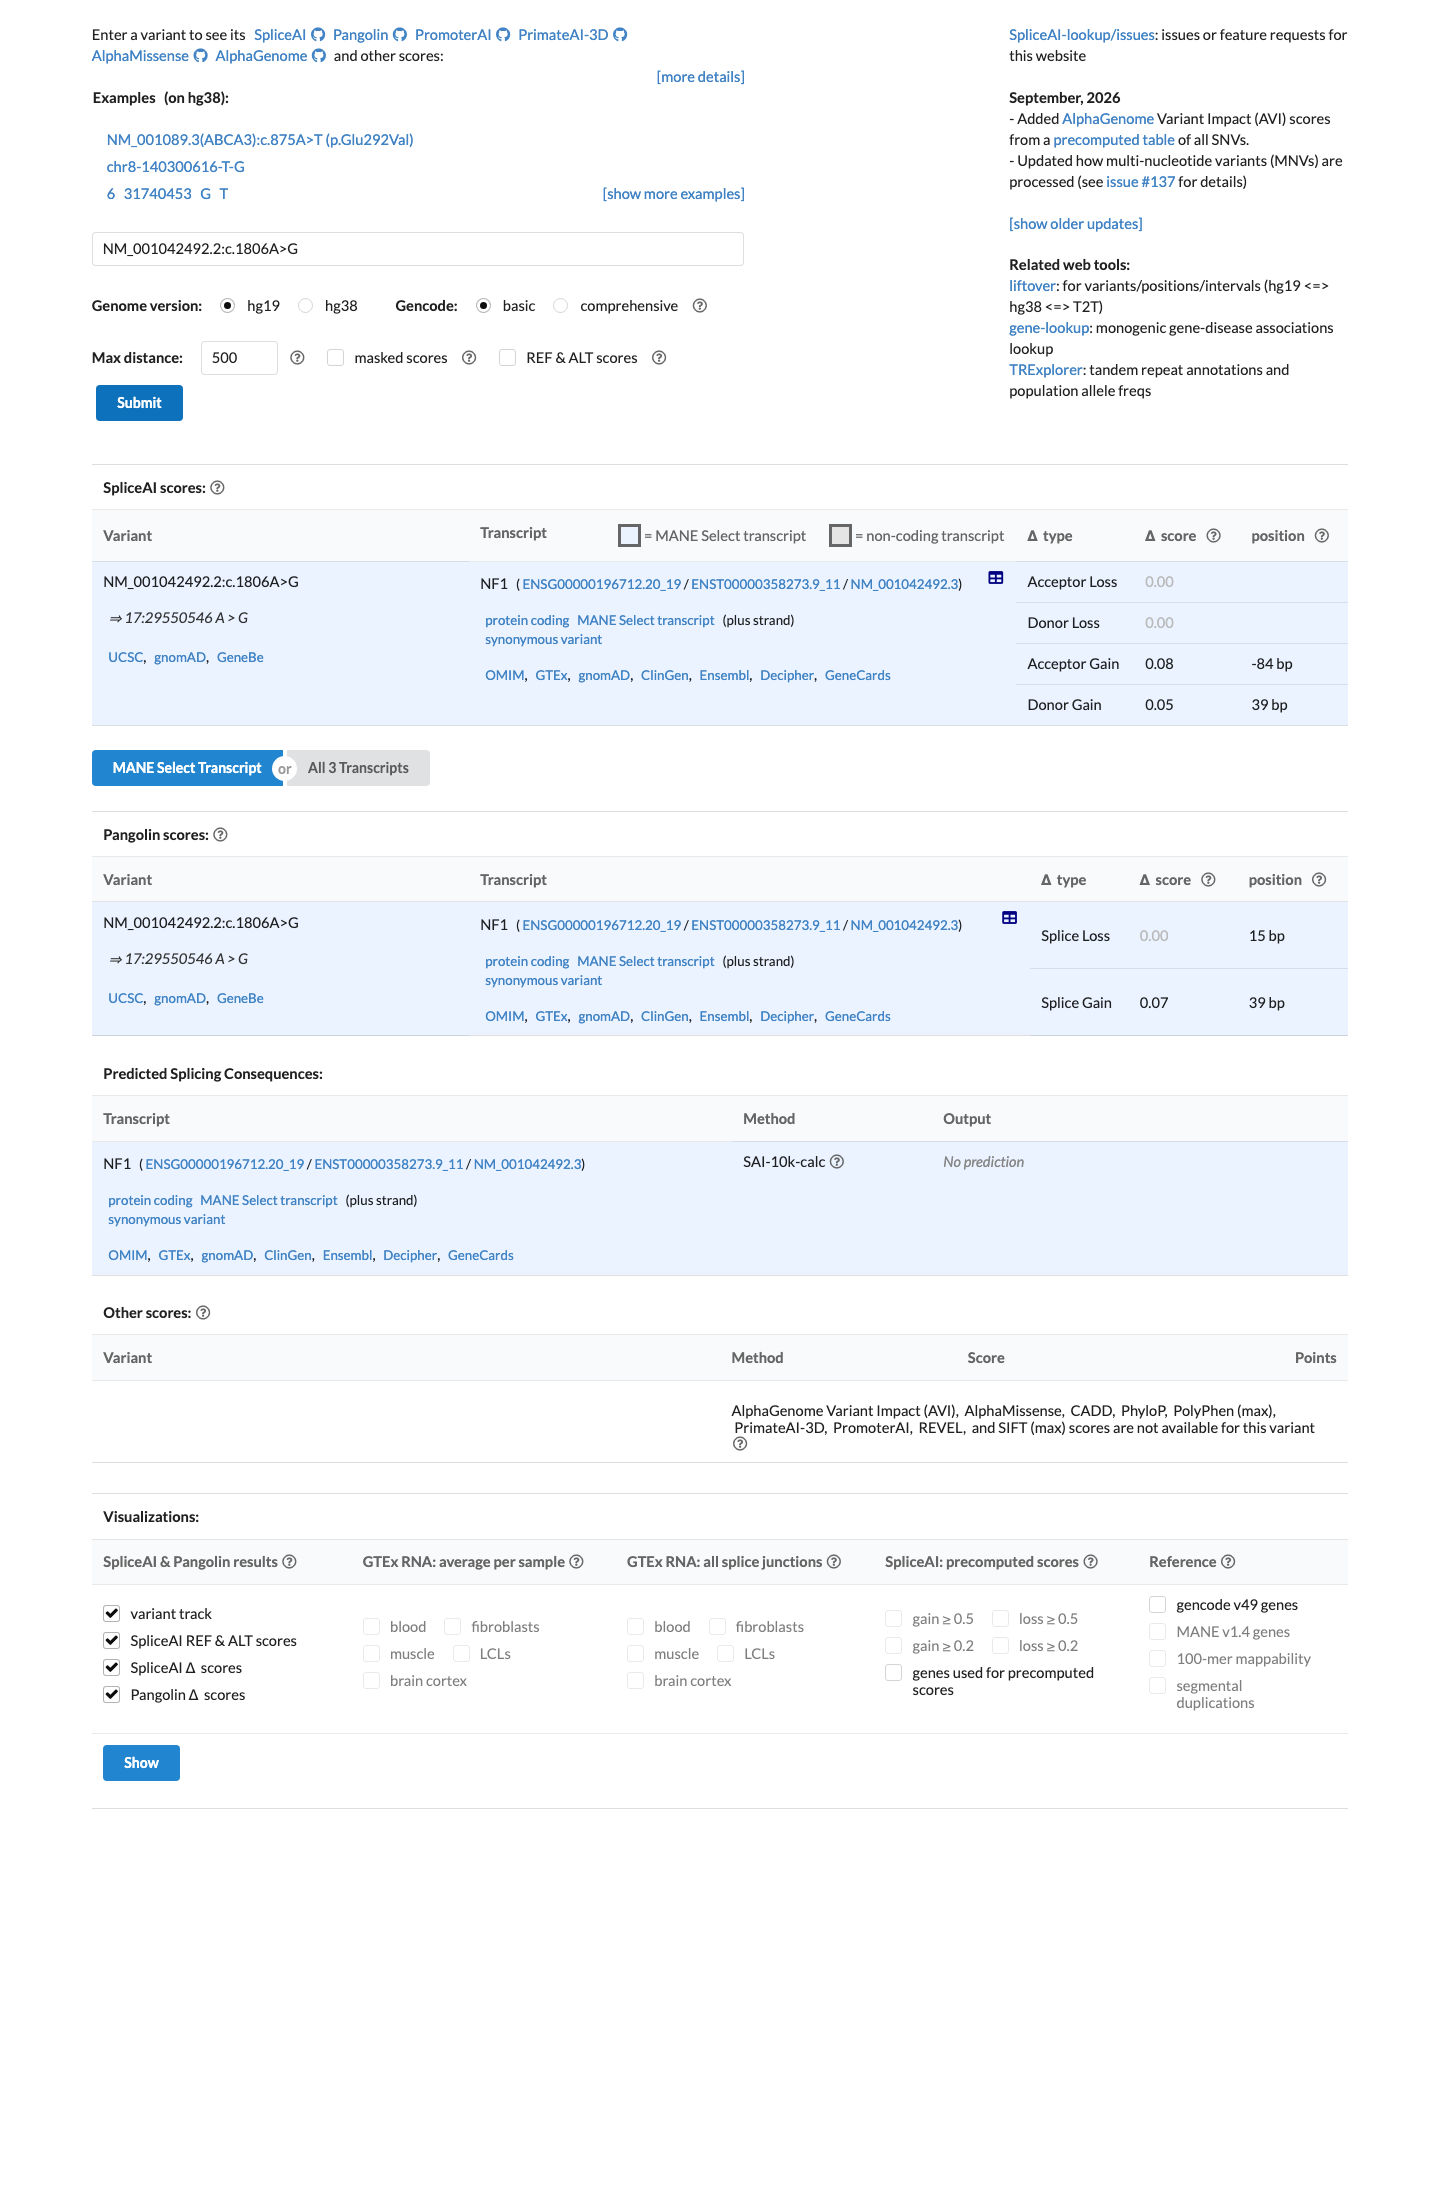
VUS: BP7 supporting is met because synonymous p.Glu602= has SpliceAI maximum delta 0.078, below the 0.1 cutoff.
BP7→VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria
may be applied incorrectly, sources may be misread, and a confident-looking
classification can still be wrong. Double-check every criterion and
its underlying evidence before relying on any call.
Gene diagram
· NM_001042492.2 · variants mapped to exon structure
NF1NM_001042492.2
Fetching transcript structure from UCSC…
Exons
—
Transcript span
—
Strand
—
Variants mapped
—
Source
UCSC ncbiRefSeqCurated
All variants in NF1—click a row to locate it on the plot · use the link column to open its page
Protein
Location
Classification
Link
Applied criteria · 1 applied · 16 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 1
Strength Supporting Moderate Strong Very strong
✓
BP7supportingBenign
Met, supporting: synonymous p.Glu602= with SpliceAI maximum delta 0.078 below the <=0.1 no-significant-splice-impact cutoff.
The case variant is synonymous: NM_001042492.2:c.1806A>G, NP_001035957.1:p.(Glu602=).SpliceAI reports DS_AG 0.078, DS_DG 0.047, DS_AL 0.0, DS_DL 0.0, with maximum delta score 0.078.The supplied generic SpliceAI calibration defines <=0.1 as the recommended no-significant-splice-impact threshold for benign computational evidence; this calibration is attributed to Jaganathan et al. 2019 (PMID:30661751).
Assessed · not applied
· 5 not met · 11 not assessed
Pathogenic
PS2Not assessed: no documented parental genotypes or confirmed de novo occurrence are available for NM_001042492.2:c.1806A>G.
PS3Not assessed: no validated RNA, protein, or cellular assay directly evaluates NF1 c.1806A>G (p.Glu602=).
PS4Not assessed: no exact-variant case-control enrichment, affected-case series, or odds ratio is available for NM_001042492.2:c.1806A>G.
PM2Not met: the highest ancestry-specific allele frequency is 1.01764e-04, exceeding the generic PM2 cutoff of 0.0001.
PM6Not assessed: no affected individual with this exact variant is documented as apparently de novo without parental testing.
PP1Not assessed: no affected relatives, informative meioses, or family segregation data are reported for this exact variant.
PP4Not assessed: no proband phenotype or highly specific NF1 clinical presentation is documented for this variant.
PP5Not met: ClinVar shows 0 expert-panel submissions, and the exact-variant record contains no expert-panel Pathogenic or Likely pathogenic classification.
Benign
BA1Not met: the highest observed allele frequency is 1.01764e-04, far below the generic BA1 threshold of 0.05.
BS1Not met: the highest observed allele frequency is 1.01764e-04, below the generic BS1 threshold of 0.01.
BS2Not assessed: gnomAD shows carriers but provides no documented unaffected adult status or phenotype assessment to satisfy BS2.
BS3Not assessed: no validated assay demonstrates normal NF1 function for c.1806A>G (p.Glu602=).
BS4Not assessed: no confirmed variant-positive unaffected relatives with reliable age-appropriate phenotyping are documented.
BP2Not assessed: no affected-proband co-occurrence or cis/trans phase data with a pathogenic variant are documented for c.1806A>G.
BP5Not assessed: no affected individual with this variant and a separate pathogenic explanation, or BP5-specific likelihood ratio, is documented.
BP6Not met: ClinVar shows 0 expert-panel submissions, so non-expert Benign and Likely benign assertions cannot trigger BP6.
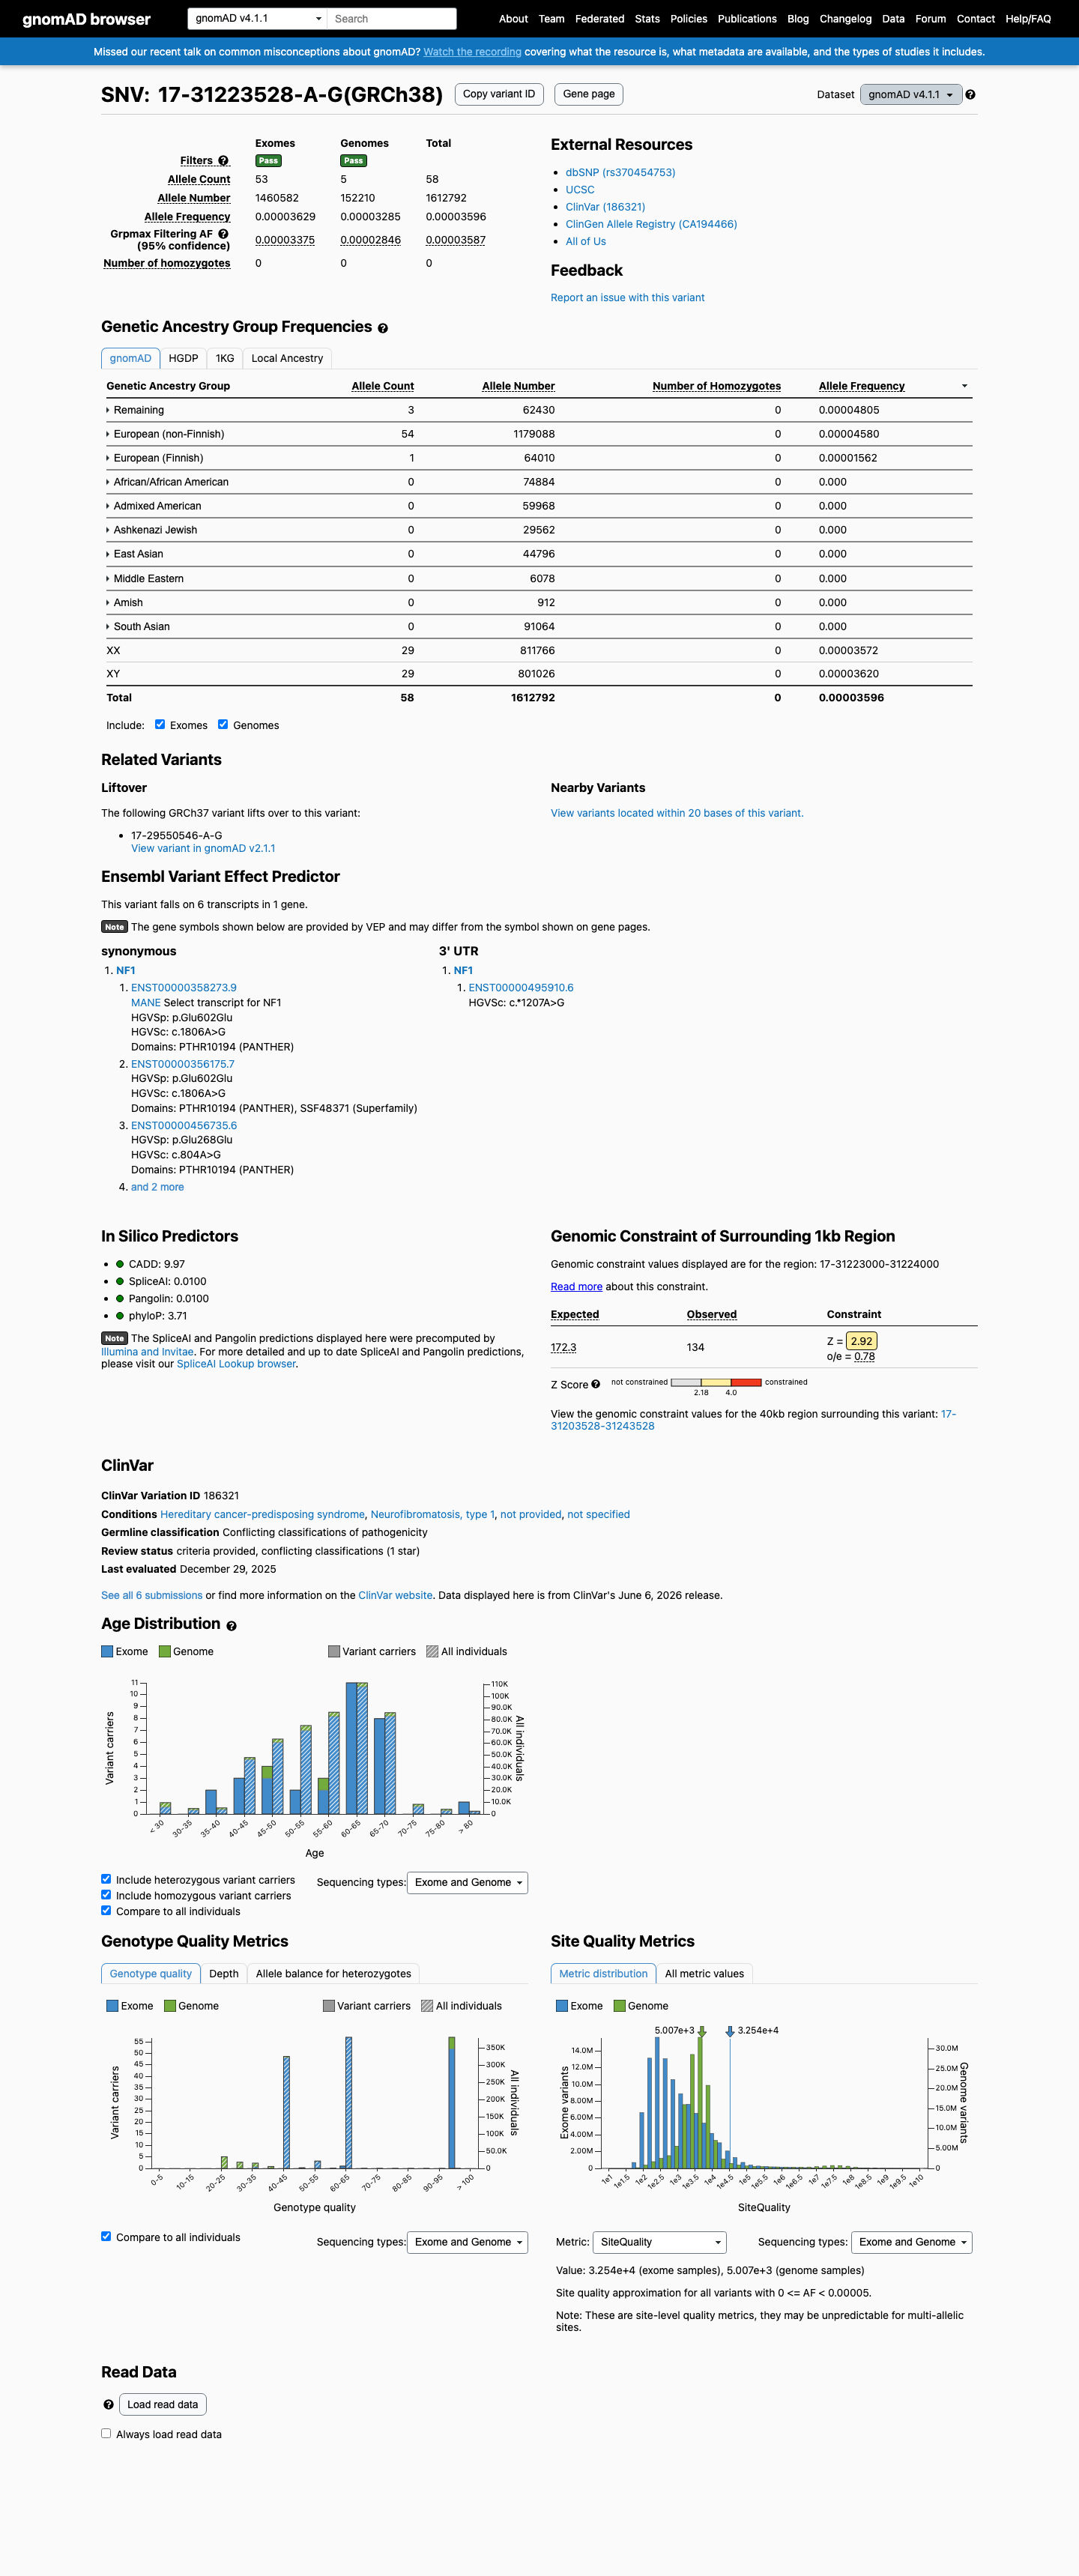
This variant is present in gnomAD v4.1 (AF= 3.59625e-05; MAF= 0.00360%, 58/1612792 alleles, homozygotes = 0) and has highest observed frequency in the Remaining individuals population (AF= 4.80538e-05; MAF= 0.00481%, 3/62430 alleles, homozygotes = 0); grpmax FAF= 3.587e-05.
v2.1
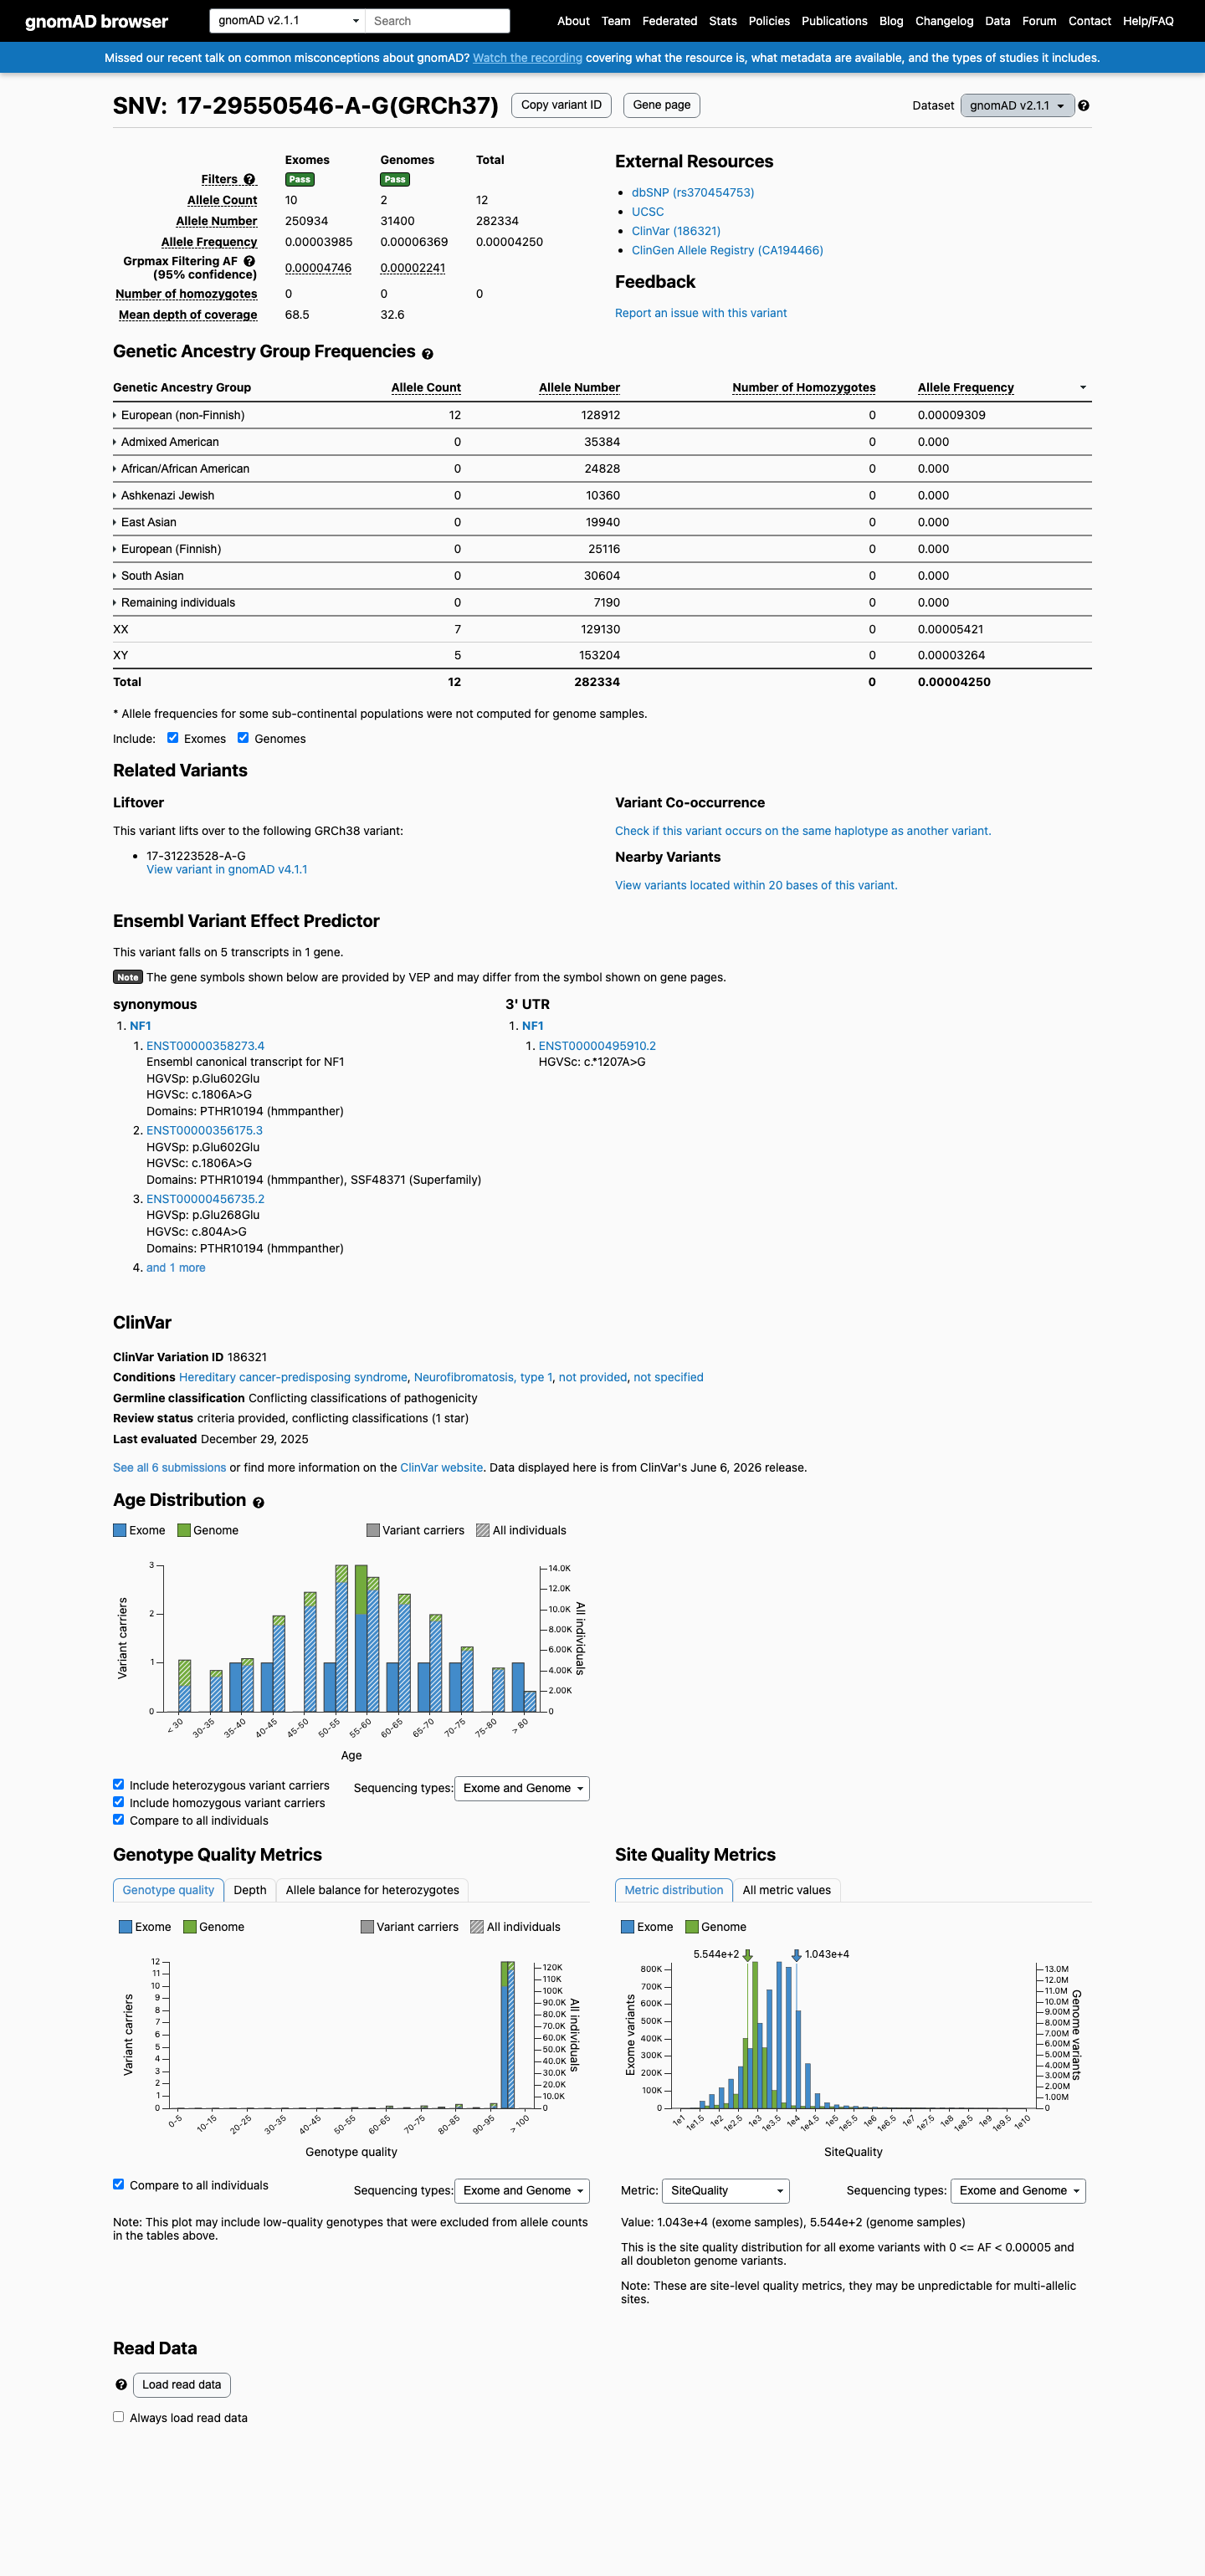
This variant is present in gnomAD v2.1 (AF= 4.25029e-05; MAF= 0.00425%, 12/282334 alleles, homozygotes = 0) and has highest observed frequency in the European (non-Finnish) population (AF= 9.30868e-05; MAF= 0.00931%, 12/128912 alleles, homozygotes = 0); grpmax FAF= 4.746e-05.
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.0036%
· 58 / 1,612,792
0 hom · FAF 0.0036%
Remaining individuals
3 / 62,430
0.0048%
European (non-Finnish)
54 / 1,179,088
0.0046%
European (Finnish)
1 / 64,010
0.0016%
+ 7 not observed (Admixed American, Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American)
gnomAD v2.1
0.0043%
· 12 / 282,334
0 hom · FAF 0.0047%
European (non-Finnish)
12 / 128,912
0.0093%
+ 7 not observed (African/African American, Admixed American, Ashkenazi Jewish, East Asian, European (Finnish), Remaining individuals, South Asian)
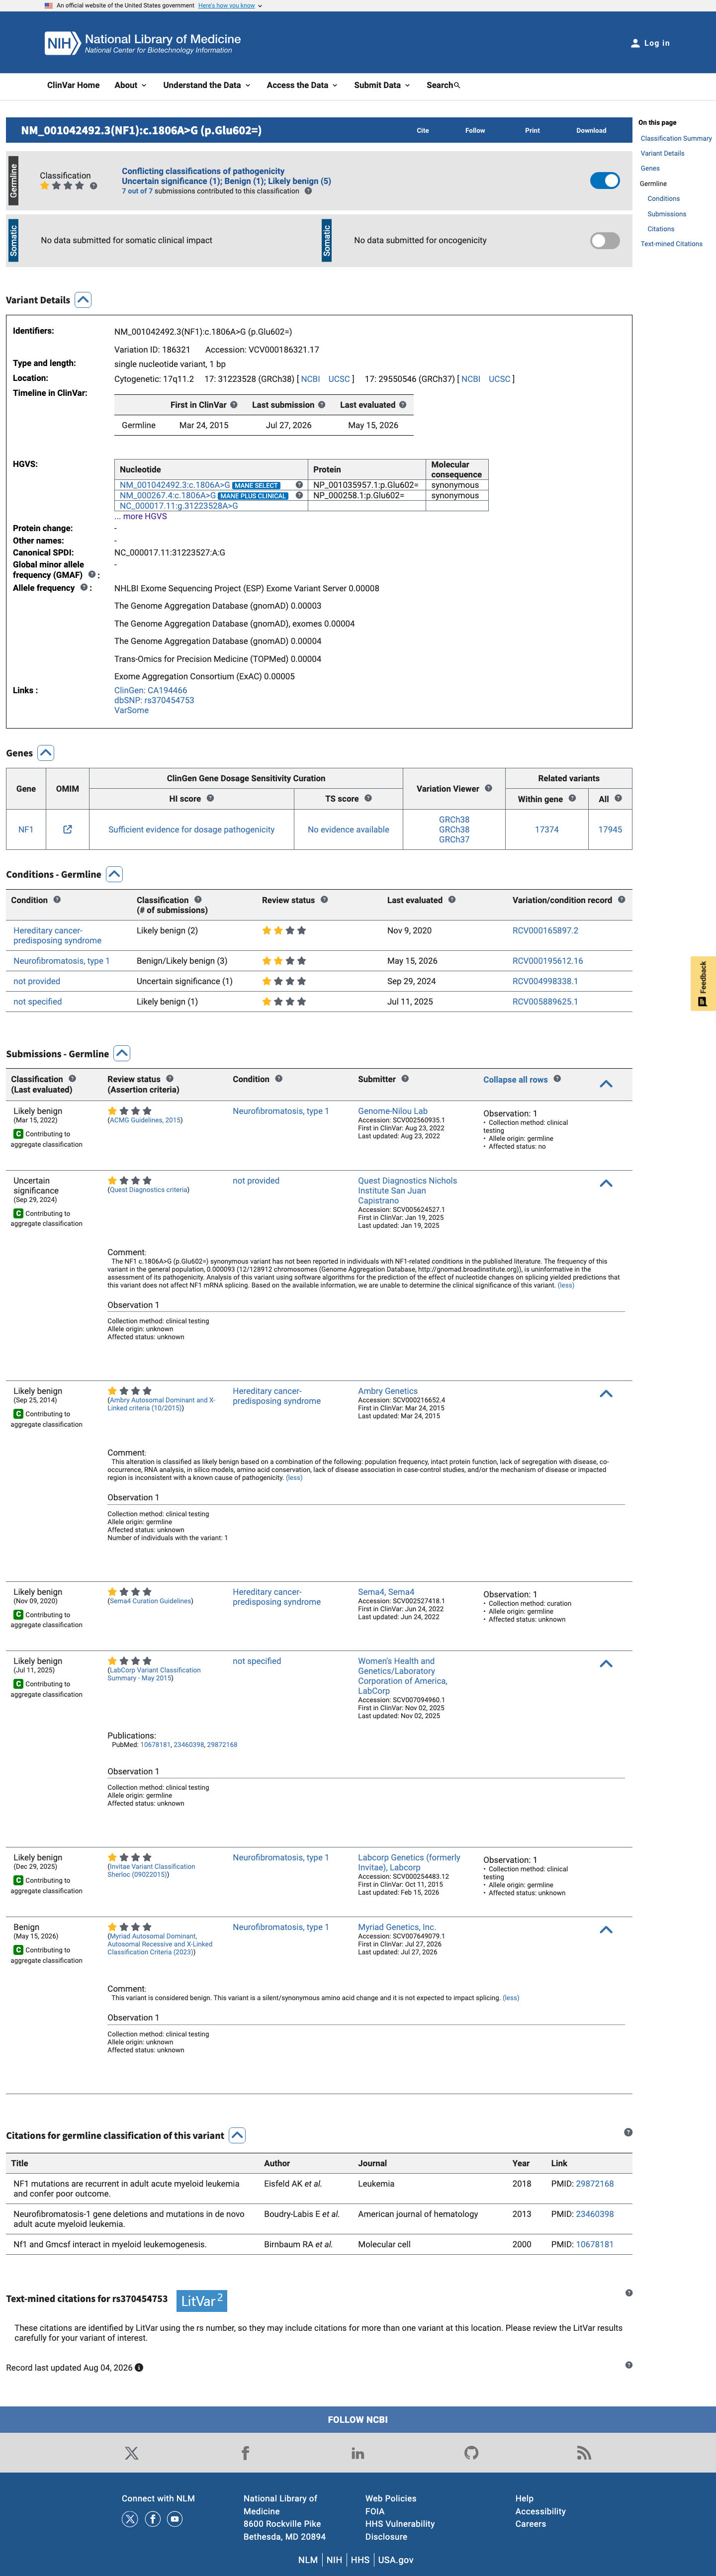
This variant has been reported in ClinVar as Likely benign (4 clinical laboratories) and as Uncertain significance (1 clinical laboratory) and as Benign (1 clinical laboratory). (ClinVarID = 186321)
Triaged references · 11 PMIDs not cited in assessment
23460398 ↗Neurofibromatosis-1 gene deletions and mutations in de novo adult acute myeloid leukemia.CLINVAR
25741868 ↗Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.CLINVAR
26467025 ↗A Standardized DNA Variant Scoring System for Pathogenicity Assessments in Mendelian Disorders.CLINVAR
29872168 ↗NF1 mutations are recurrent in adult acute myeloid leukemia and confer poor outcome.CLINVAR
10678181 ↗Nf1 and Gmcsf interact in myeloid leukemogenesis.CLINVAR
15604628 ↗Genetic cancer risk assessment and counseling: recommendations of the national society of genetic counselors.CLINVAR
25394175 ↗A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment.CLINVAR
26324357 ↗American Society of Clinical Oncology Policy Statement Update: Genetic and Genomic Testing for Cancer Susceptibility.CLINVAR
26140447 ↗Points to Consider: Ethical, Legal, and Psychosocial Implications of Genetic Testing in Children and Adolescents.CLINVAR
28492532 ↗Sherloc: a comprehensive refinement of the ACMG-AMP variant classification criteria.CLINVAR
33939658 ↗The North American Neuroendocrine Tumor Society Consensus Guidelines for Surveillance and Management of Metastatic and/or Unresectable Pheochromocytoma and Paraganglioma.CLINVAR