Back
NM_001276270.2:c.1375T>A
p.Phe459Ile · MBD4
ACMG/AMP
0%
complete
Final classification
VUS
PM2PP3
MBD4
c.1375T>A
p.Phe459Ile
missense · exon 5

MBD4 encodes a DNA glycosylase that detects and repairs deaminated methyl-cytosines in DNA, using a methyl-CpG binding domain to anchor to methylated DNA and a glycosylase domain to carry out mismatch repair in CpG-rich regions. The protein also helps repress transcription from methylated gene promoters. MBD4 acts as a tumor suppressor, and its loss has been linked to colorectal cancer, acute myeloid leukemia, and uveal melanoma.

This variant

This missense change affects MBD4, a DNA glycosylase that repairs deaminated methyl-cytosines and functions as a tumor suppressor.

Transcript
NM_001276270.2
HGVS · transcript:coding
NM_001276270.2:c.1375T>A
GRCh38
chr3:129433868 A>T
GRCh37
chr3:129152711 A>T
VUS: PM2 (supporting) and PP3 (supporting) provide two supporting criteria, which do not satisfy a generic ACMG/AMP pathogenic, benign, or likely classification threshold.
Classification rationale
PM2PP3 VUS
MBD4 c.1375T>A missense · exon 5

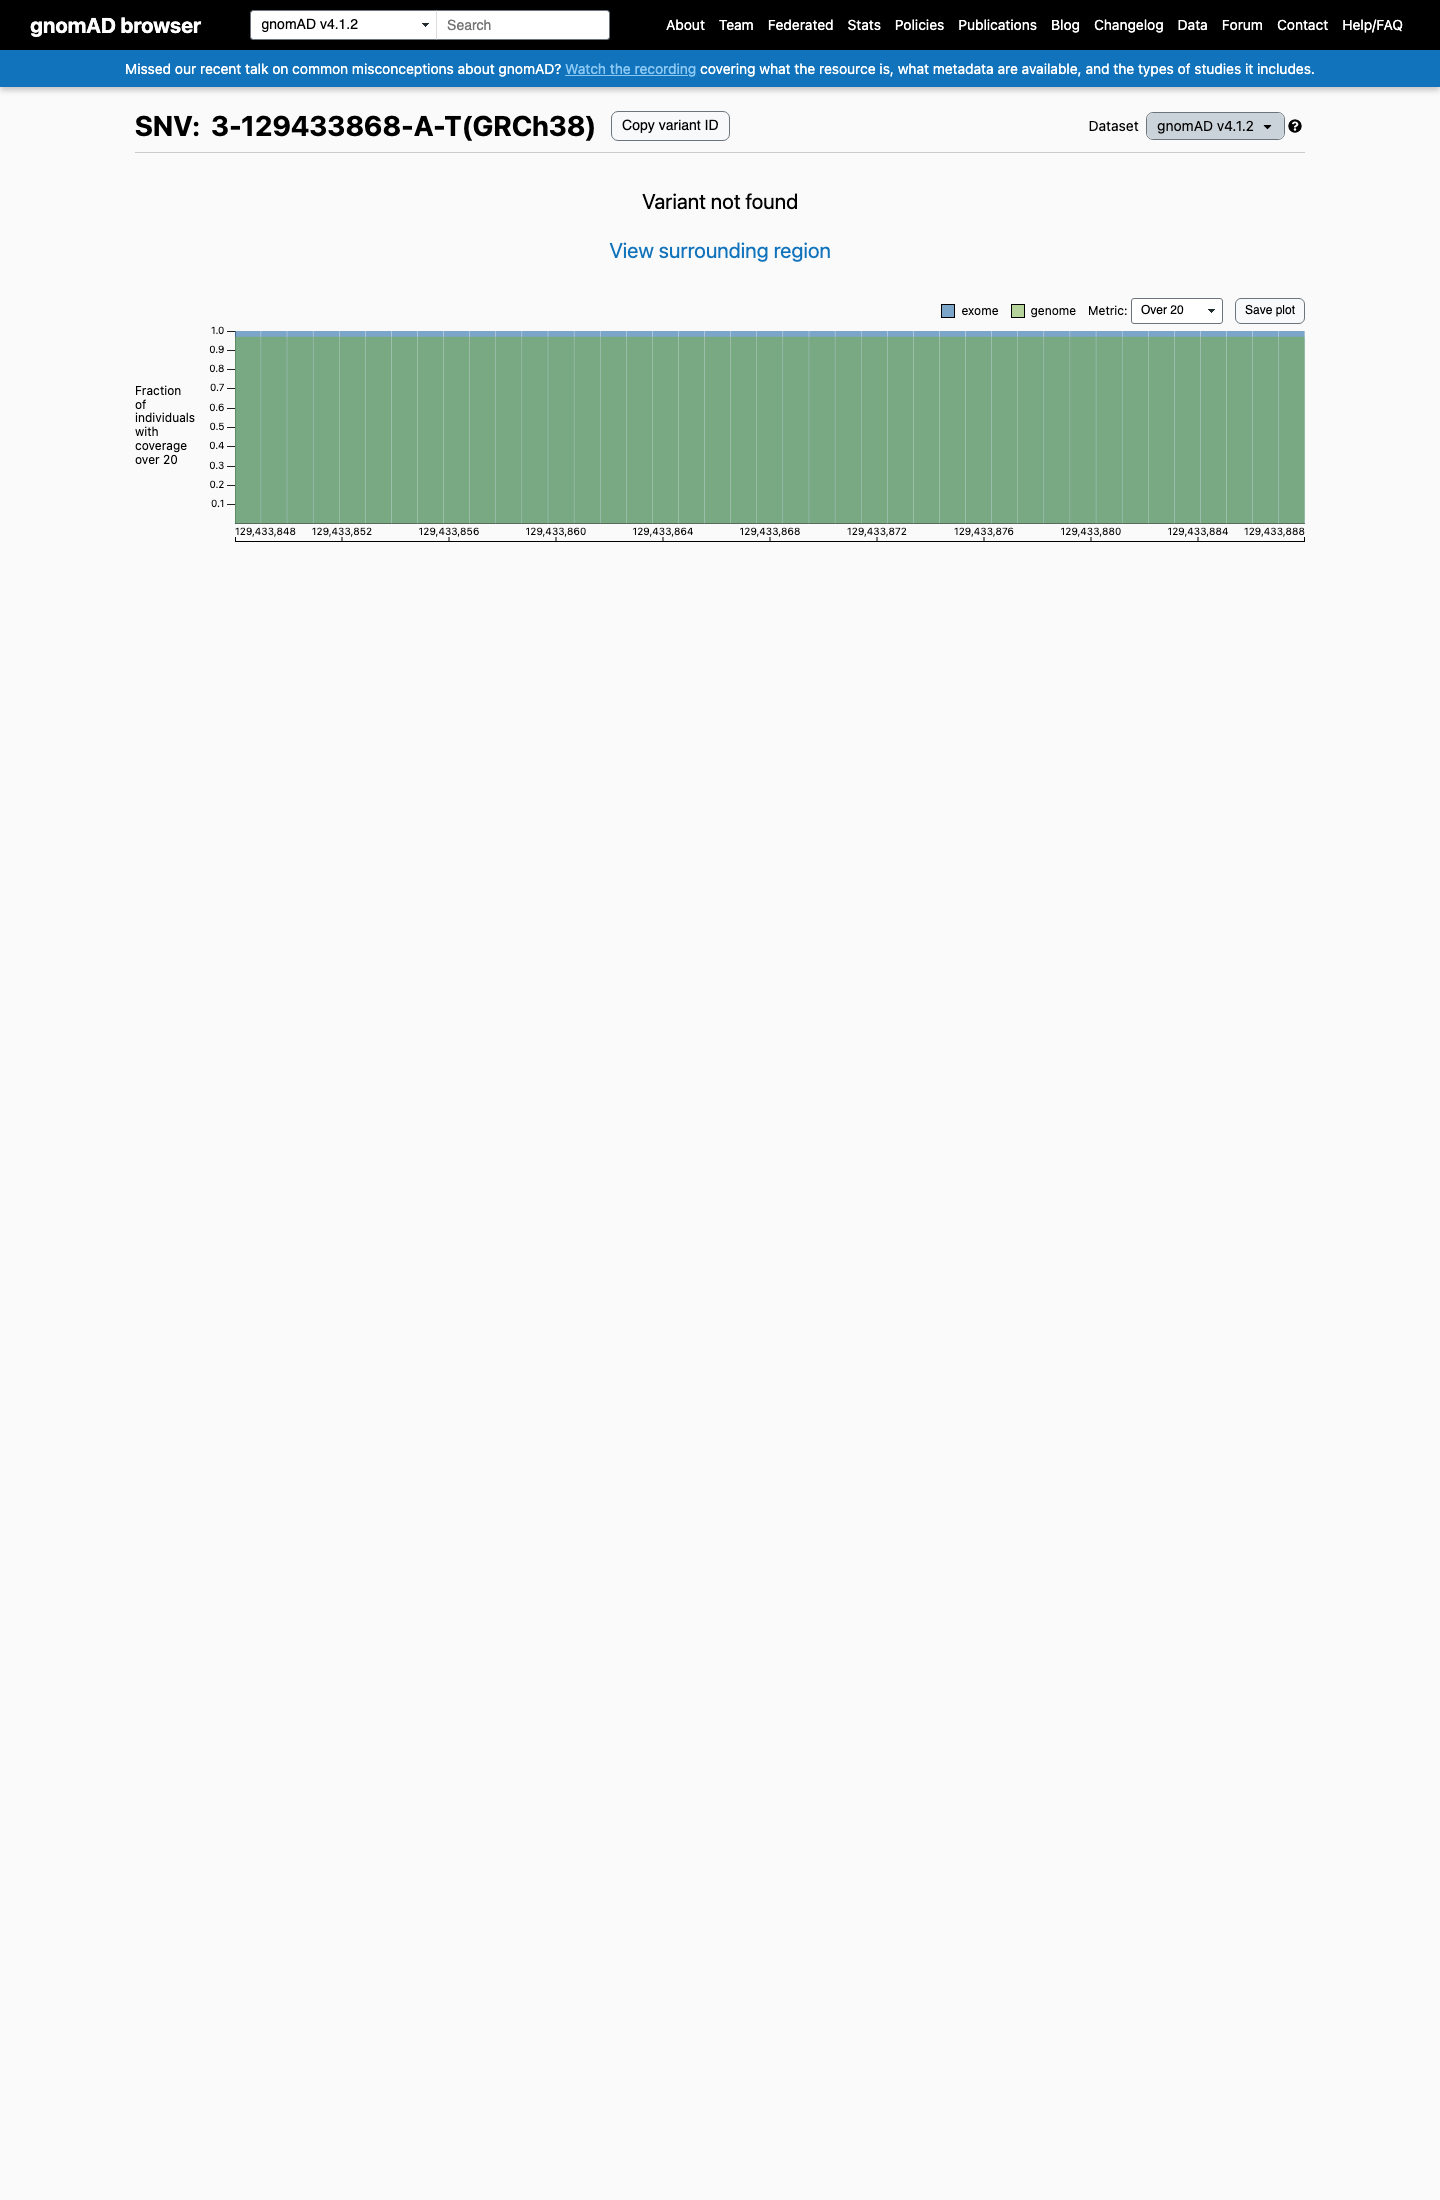
PM2 supporting: the variant is absent from gnomAD v2.1 and v4.1. PP3 supporting: REVEL 0.658 meets the calibrated missense threshold of at least 0.644.

PM2 + PP3 → VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_001276270.2 · variants mapped to exon structure
MBD4 NM_001276270.2
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 22 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
PM2 supporting Pathogenic
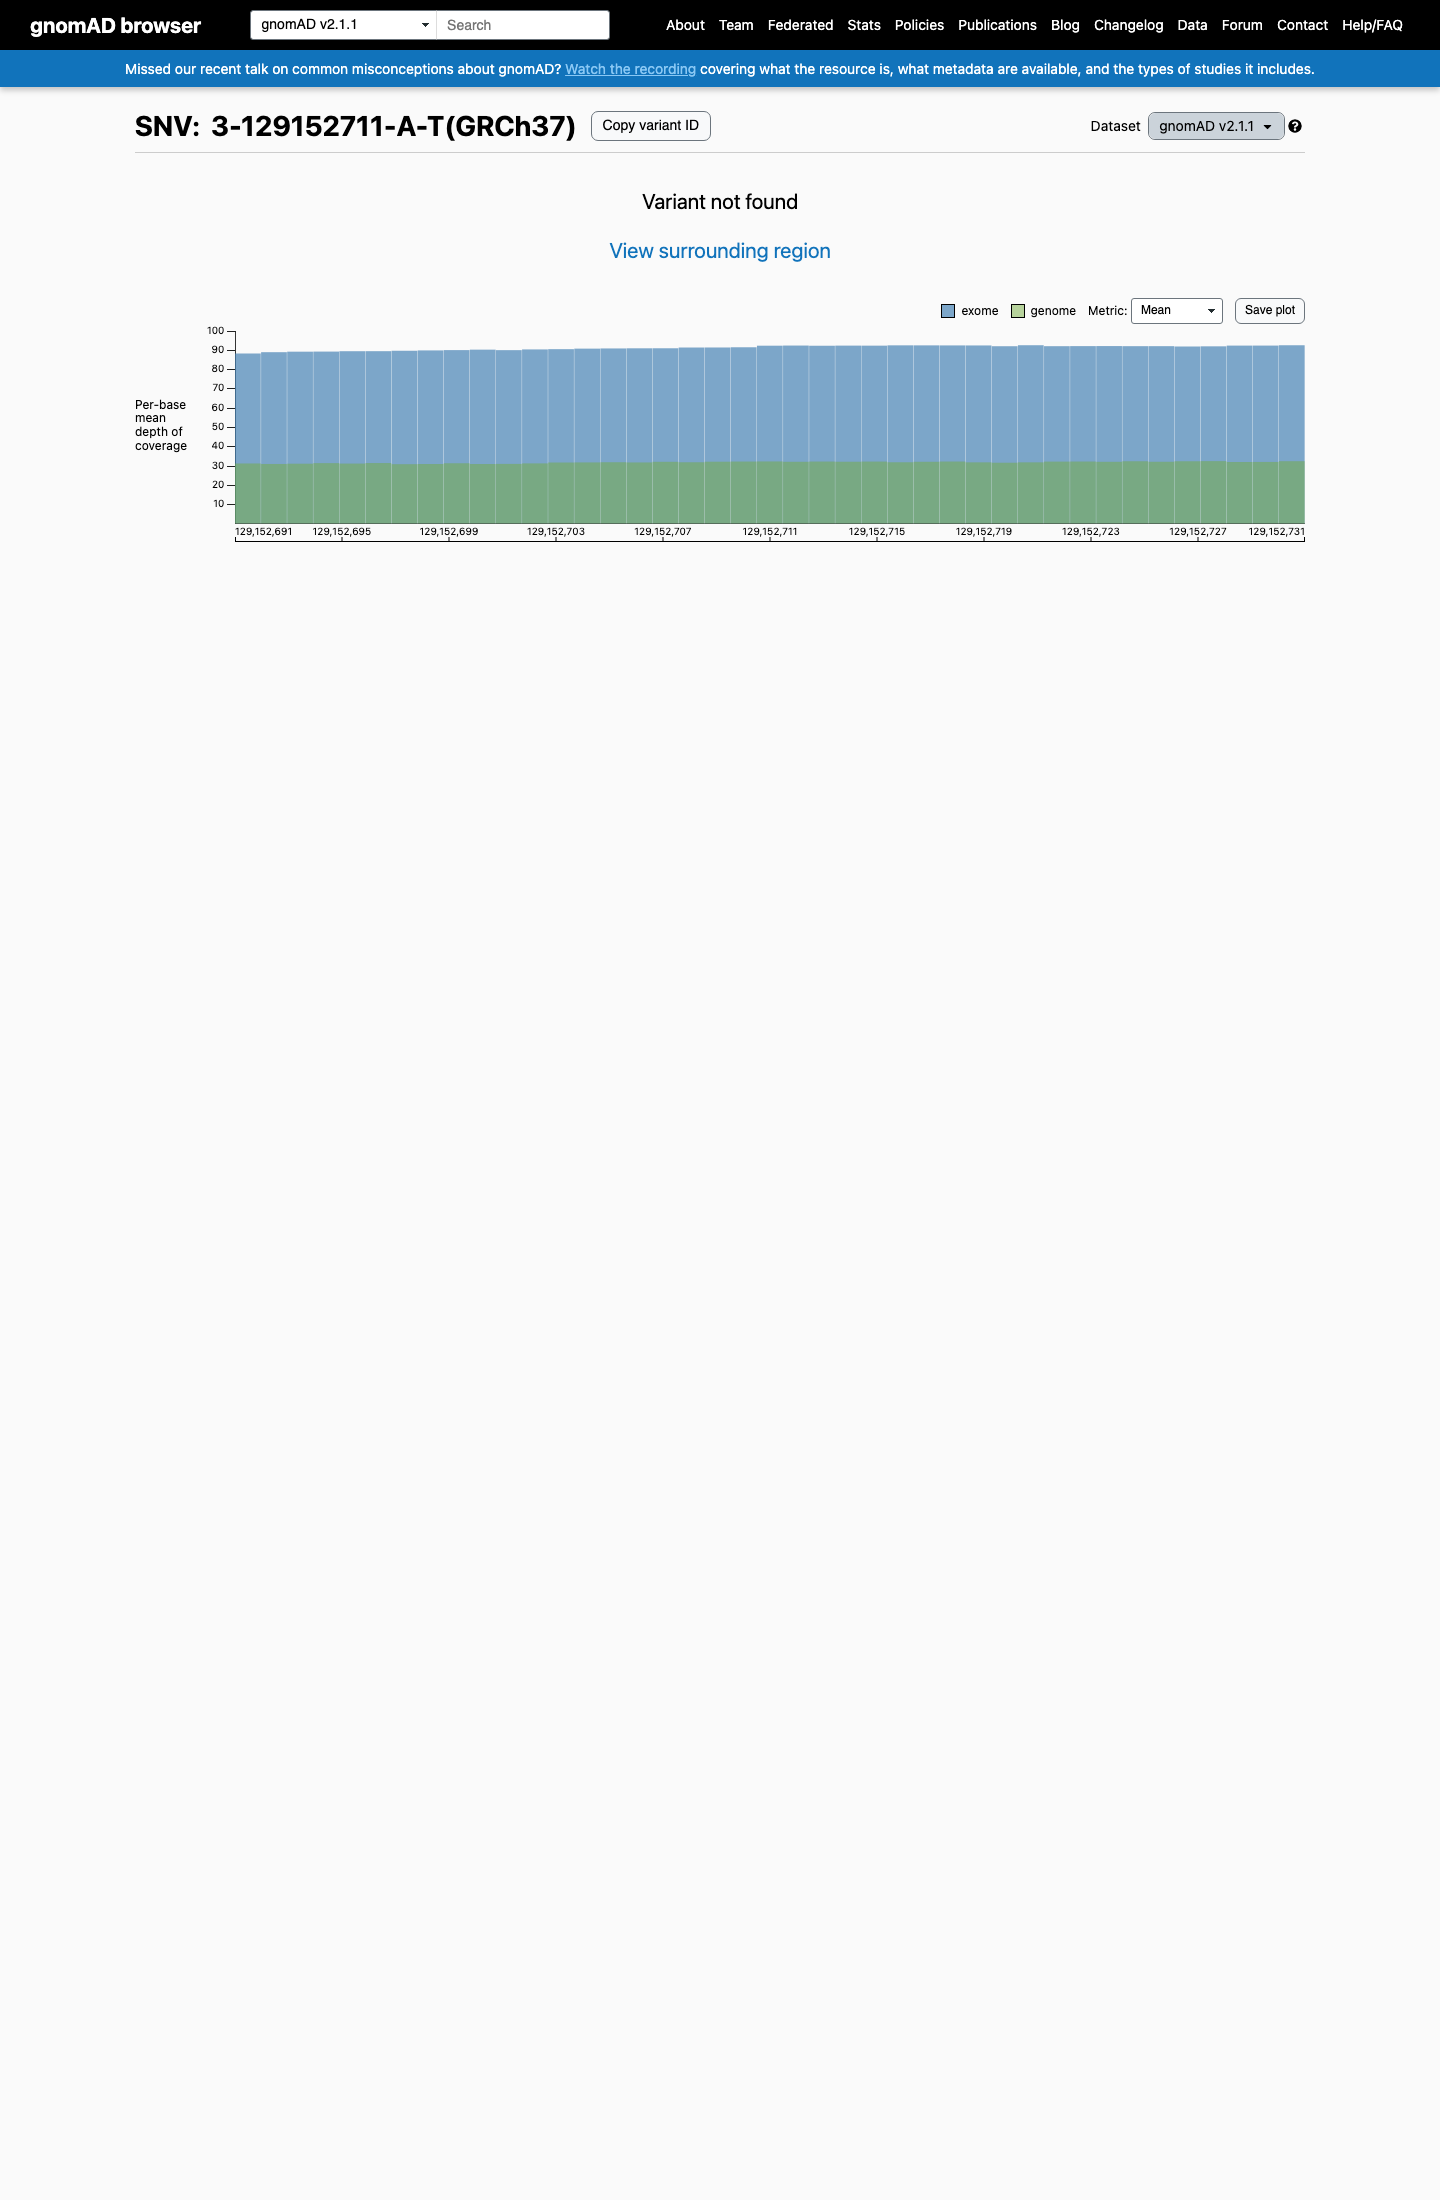
Met at supporting: the variant is absent from gnomAD v2.1 and v4.1, consistent with allele frequency <=0.0001 for generic PM2.
gnomAD v2.1 reports the variant as absent.gnomAD v4.1 reports the variant as absent.The available gnomAD v2.1 non-cancer and v3.1 non-cancer records also report the variant as absent.
PP3 supporting Pathogenic
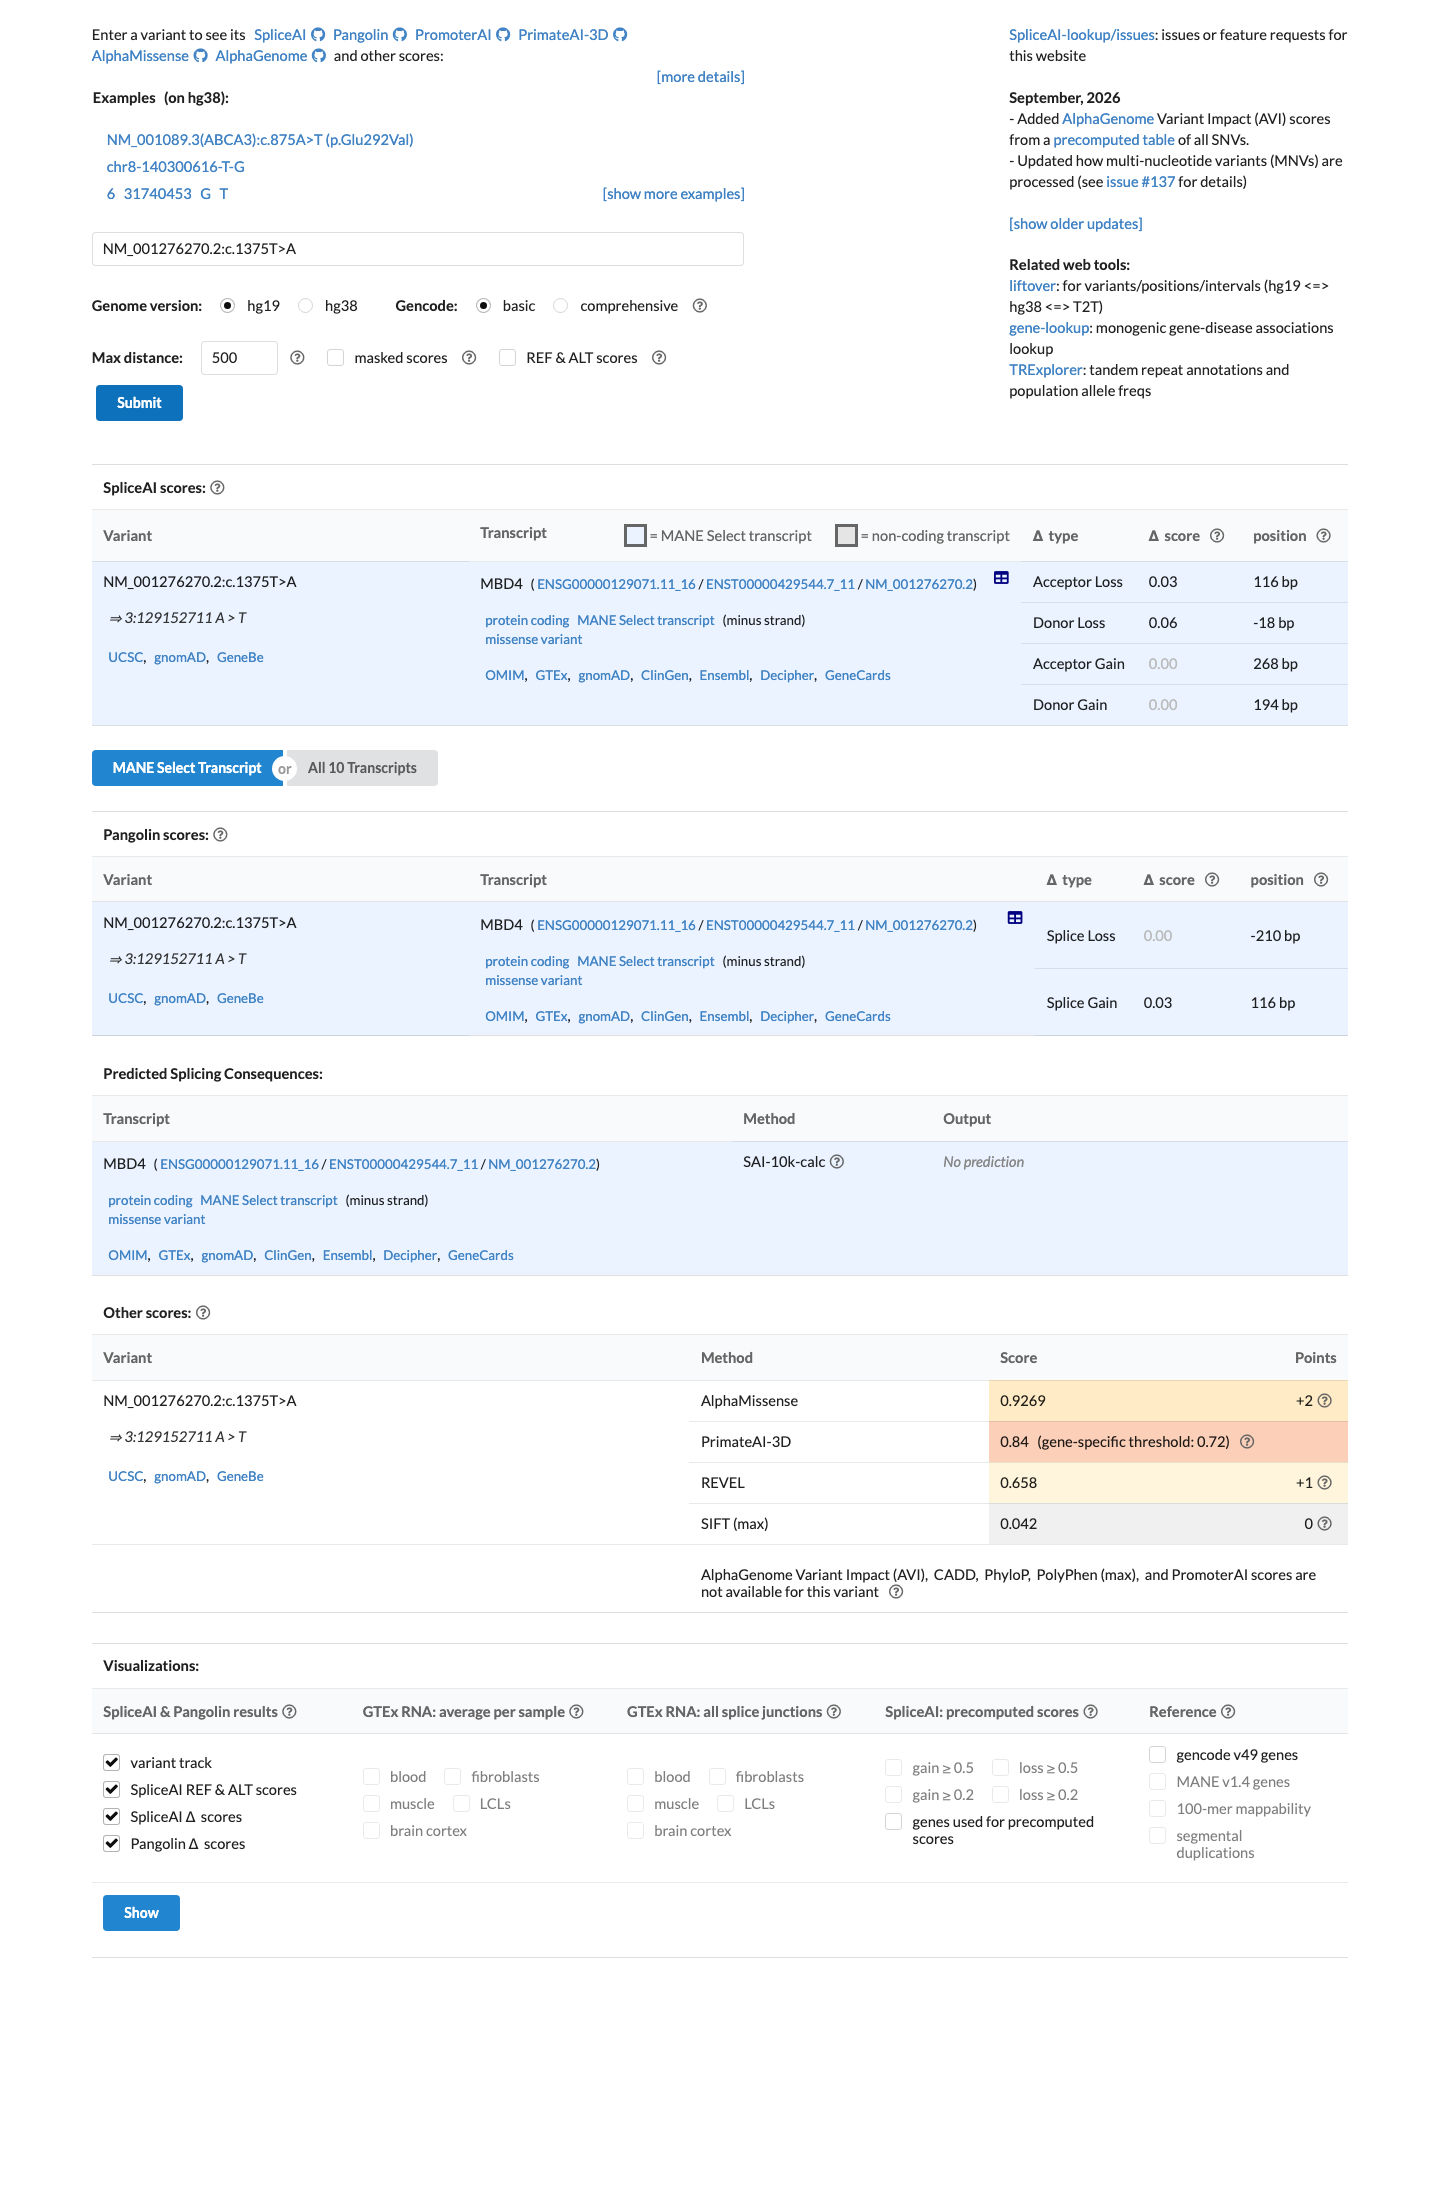
Met, supporting: REVEL 0.658 meets the >=0.644 missense PP3 threshold calibrated by ClinGen SVI (PMID:36413997).
NM_001276270.2:c.1375T>A is annotated as the missense variant p.Phe459Ile, making REVEL the applicable PP3/BP4 path.The recorded REVEL score is 0.658, which meets the generic ClinGen SVI supporting PP3 threshold of >=0.644; threshold source: Pejaver et al. 2022, PMID:36413997.SpliceAI was not evaluated for PP3 because the variant is missense and the criterion requires one consequence-matched predictor path; BayesDel was not used because no generic ACMG strength calibration is established.
Assessed · not applied · 6 not met · 16 not assessed
Pathogenic
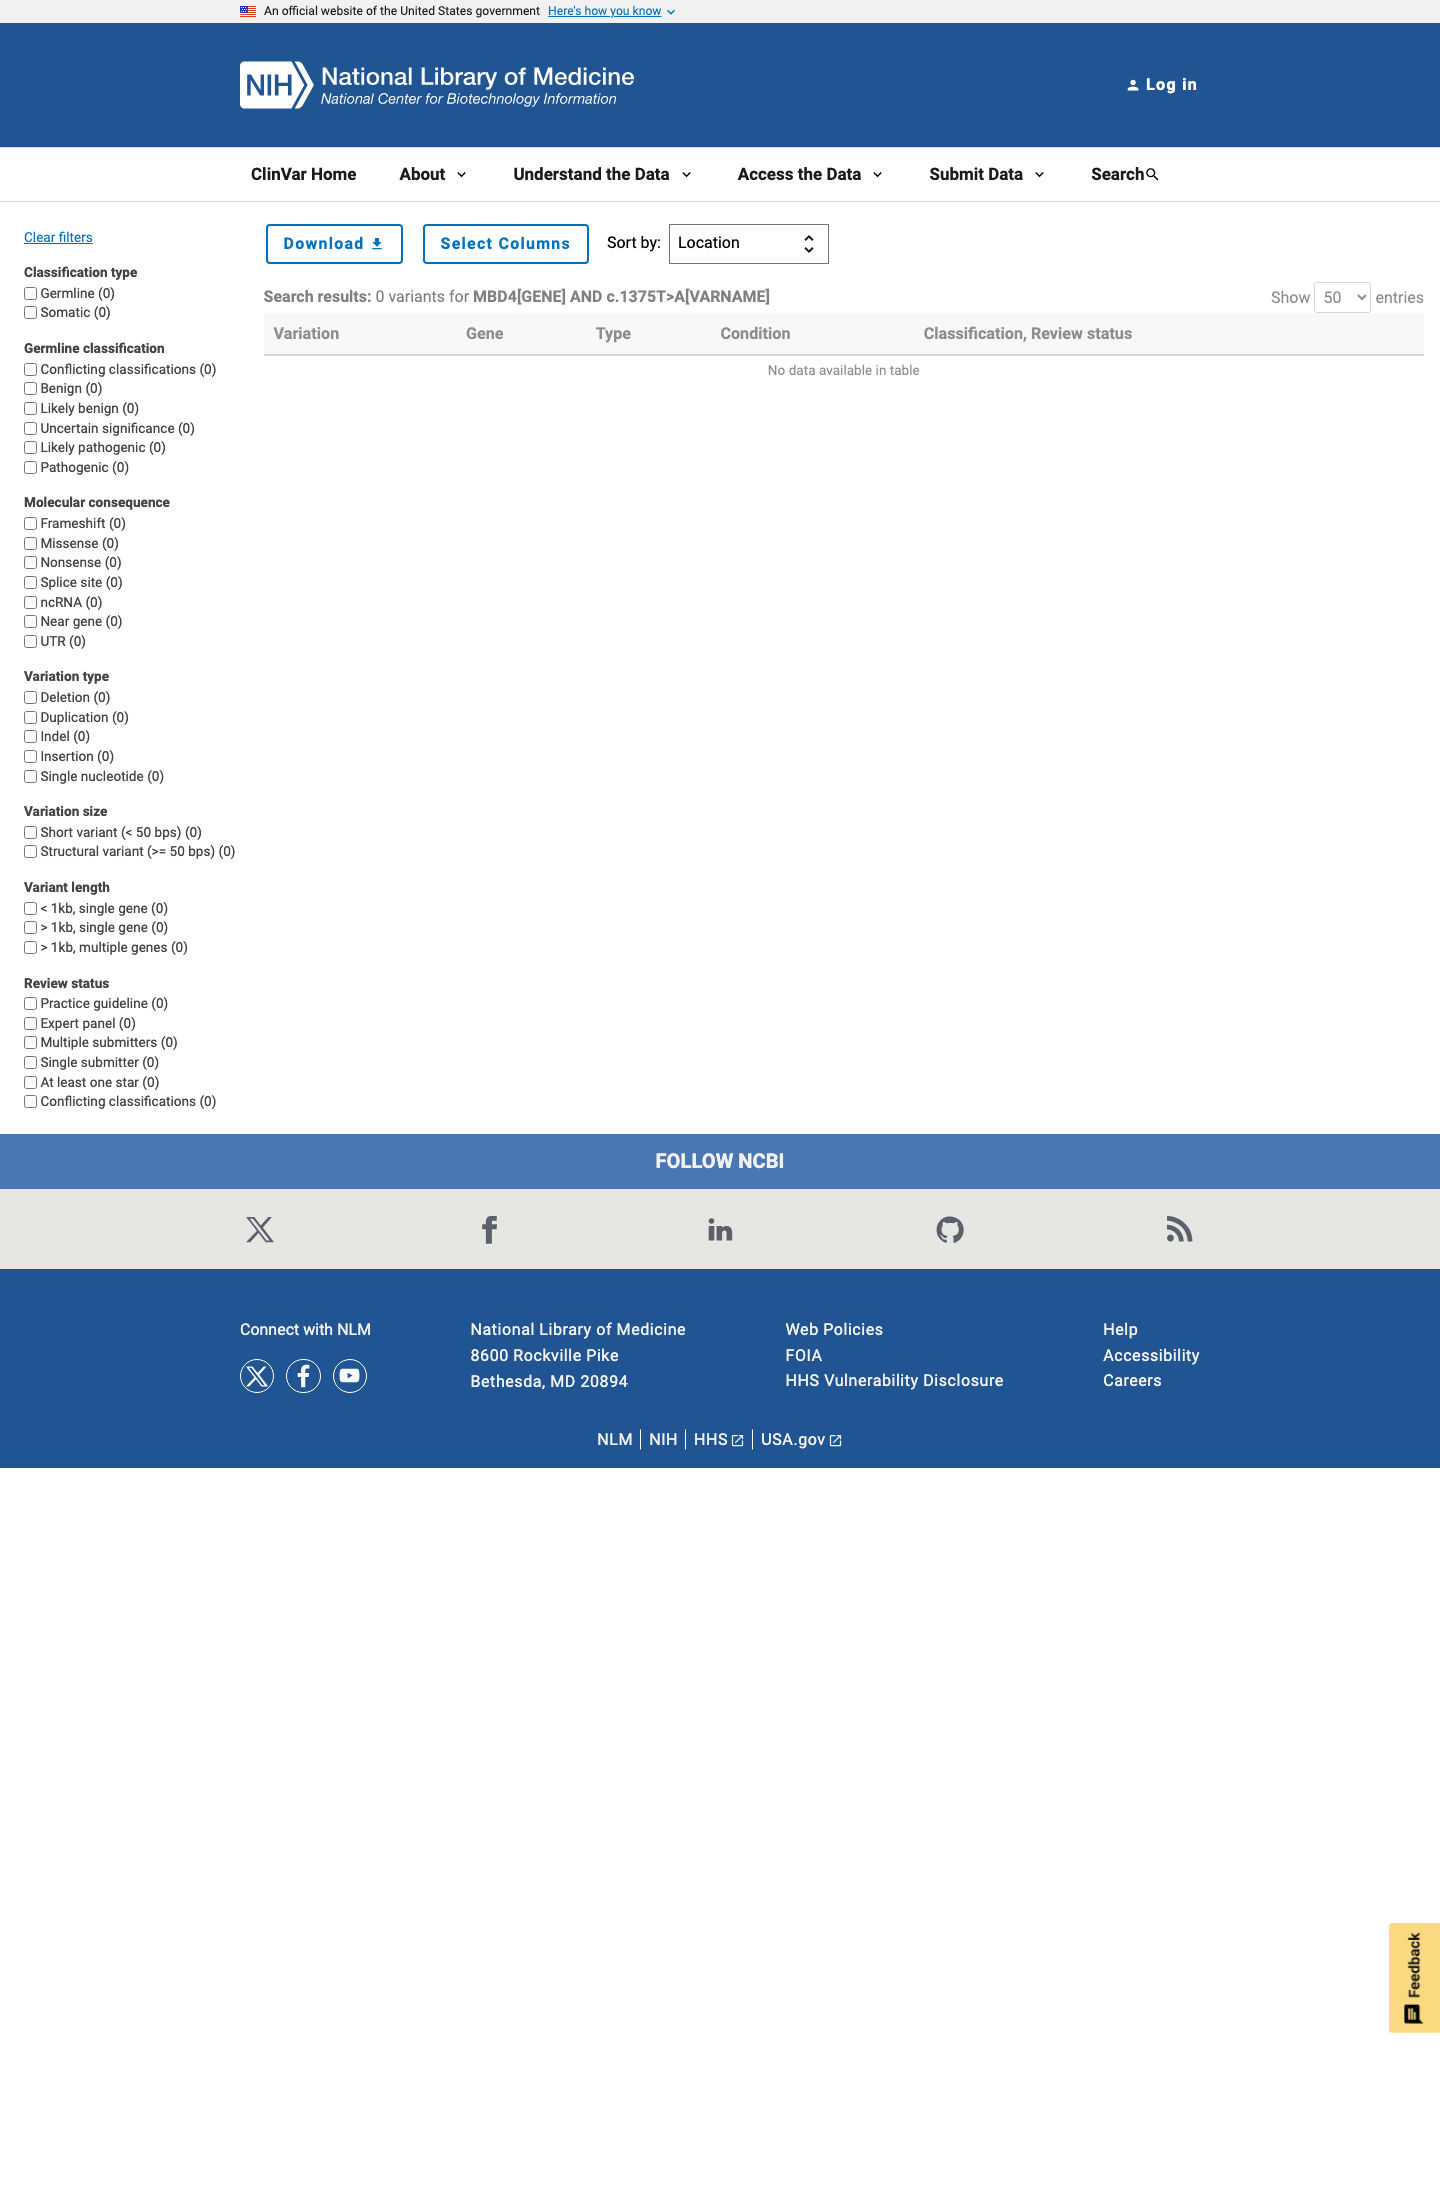
PS1 Not assessed: no established pathogenic p.Phe459Ile comparator was identified, and ClinVar returned no record for c.1375T>A.
PS2 Not assessed: no parental genotypes or confirmed absence of c.1375T>A in both parents are documented for de novo evaluation.
PS3 Not assessed: No variant-specific functional assay, validated controls, or quantitative activity result was available for MBD4 p.Phe459Ile.
PS4 Not assessed: no affected-carrier, control-comparison, or case-control enrichment data are available for this variant.
PM1 Not assessed: no authoritative MBD4 domain table covered residue F459, and the hotspot search returned no statistically significant hotspot.
PM3 Not assessed: no affected-proband observation, second pathogenic allele, phase result, or inheritance data is available for MBD4 p.Phe459Ile.
PM5 Not assessed: zero eligible pathogenic or likely pathogenic alternate missense comparators were identified at MBD4 residue 459.
PM6 Not assessed: no reported suspected de novo occurrence with phenotype and proband evidence is available when parental testing is absent.
PP1 Not assessed: no informative affected-relative segregation, unaffected-relative testing, meioses, or phenotype concordance is documented.
PP2 Not met: available MBD4 evidence supports loss of function, not the required predominantly missense disease mechanism with uncommon benign missense variation.
PP4 Not assessed: no individual-level phenotype or highly specific clinical diagnosis is supplied for the variant carrier.
PP5 Not met: ClinVar reports no exact-variant result, so no expert-panel Pathogenic or Likely pathogenic assertion supports PP5.
Benign
BA1 Not met: the variant is absent from gnomAD v2.1 and v4.1, far below the generic BA1 allele-frequency threshold of >=0.05.
BS1 Not met: the variant is absent from gnomAD v2.1 and v4.1, with no observed allele frequency reaching the generic BS1 threshold of >=0.01.
BS2 Not assessed: gnomAD shows no variant observation, but provides no unaffected adult homozygote or healthy-carrier observation for BS2.
BS3 Not assessed: No controlled functional assay demonstrated normal MBD4 activity for p.Phe459Ile.
BS4 Not assessed: no appropriately tested unaffected relatives carrying c.1375T>A or other informative non-segregation evidence is documented.
BP1 Not assessed: MBD4 loss-of-function evidence exists, but a primarily truncating disease variant spectrum has not been established.
BP2 Not assessed: no pathogenic comparator variant, cis/trans phase result, affected-status context, or inheritance data is available for MBD4 p.Phe459Ile.
BP4 Not met: REVEL 0.658 exceeds the <=0.29 missense BP4 threshold calibrated by ClinGen SVI (PMID:36413997).
BP5 Not assessed: no patient-level alternative molecular diagnosis is documented to explain the relevant phenotype.
BP6 Not met: ClinVar reports no exact-variant result, so no expert-panel Benign or Likely benign assertion supports BP6.
N/A · 4 PVS1 · PM4 · BP3 · BP7
Research & evidence
Population frequency
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
Absent from gnomAD v4.1.
v2.1
Absent from gnomAD v2.1.
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
gnomAD v2.1
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
ClinVar screenshot
ClinVar
This variant is absent from ClinVar.
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.06). REVEL score = 0.658. BayesDel score = 0.186614.
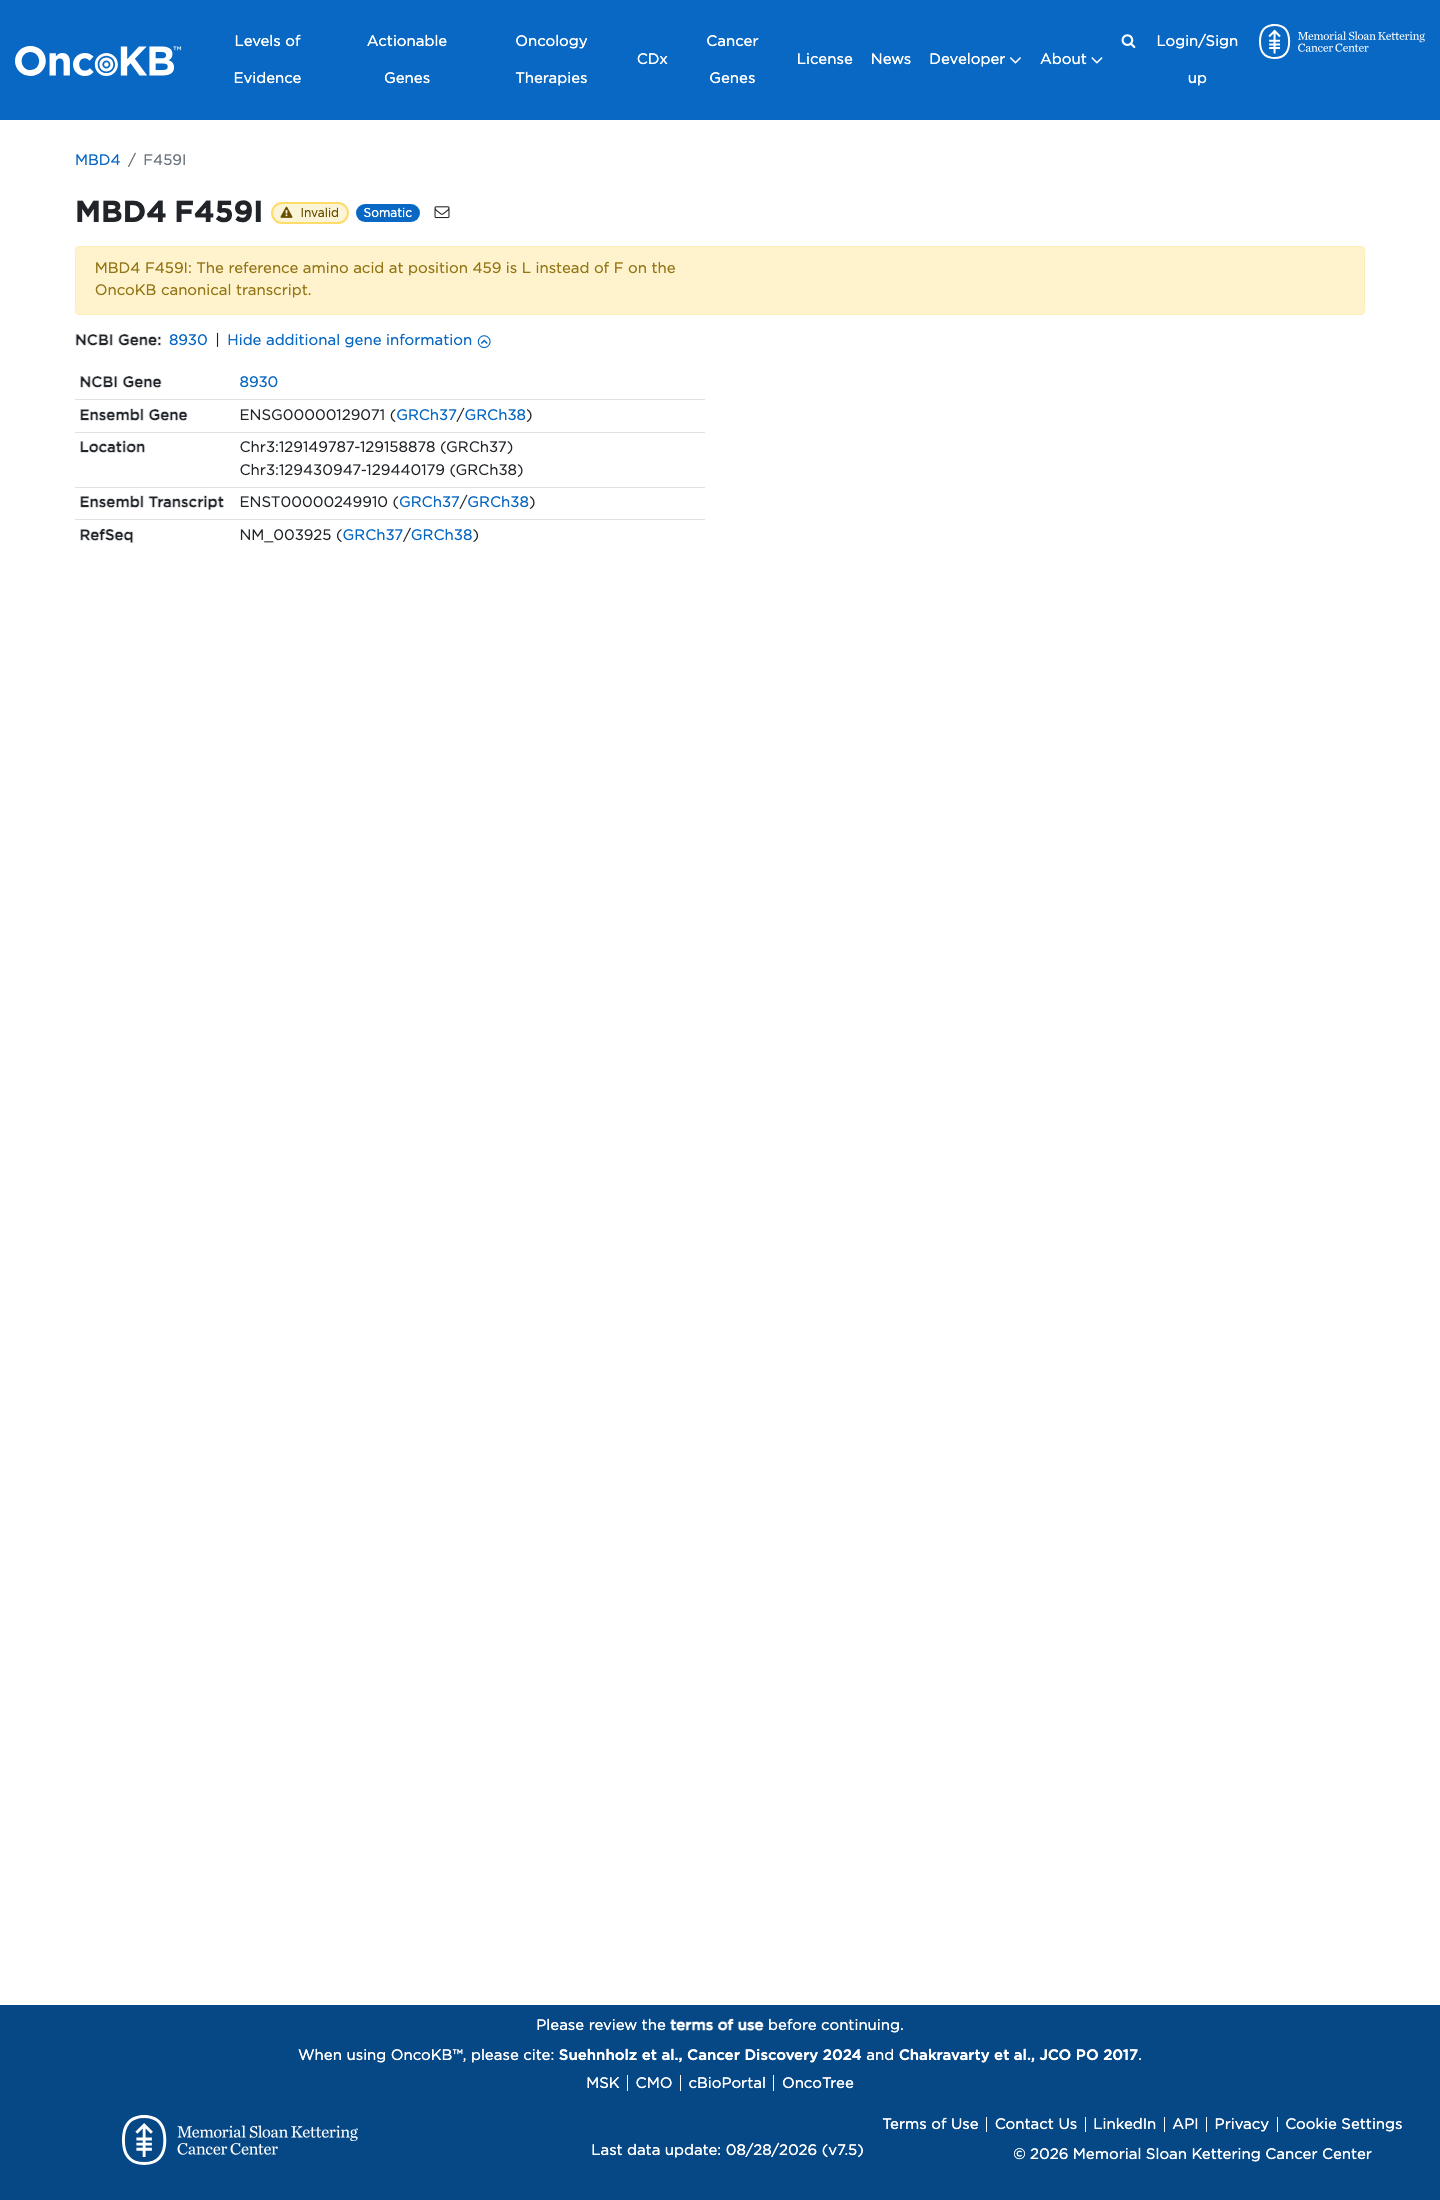
Functional / OncoKB screenshot
Functional
OncoKB identified curated literature and non-variant-specific oncogenicity context for review; listed oncogenicity label: not classified.
OncoKB ↗
COSMIC screenshot
COSMIC
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots