Back
NM_004655.4:c.203G>A
p.Arg68Gln · AXIN2
ACMG/AMP
0%
complete
Final classification
VUS
BP1BP4
AXIN2
c.203G>A
p.Arg68Gln
missense · exon 2

AXIN2 is a scaffolding protein that helps regulate the Wnt signaling pathway, a key developmental pathway controlling cell growth and fate. It is part of the beta-catenin destruction complex, where it helps mark beta-catenin for degradation when Wnt signaling is off, and it also assists in relaying Wnt signals to the nucleus when the pathway is active. AXIN2 acts as a tumor suppressor, and mutations in the gene have been linked to colorectal cancer as well as familial tooth agenesis with predisposition to colorectal cancer. Its expression has also been associated with prostate cancer recurrence.

This variant

AXIN2 c.203G>A (p.Arg68Gln) alters a residue in the tankyrase-binding region of a Wnt/beta-catenin destruction-complex scaffold and tumour suppressor whose inherited loss of function is associated with colorectal cancer predisposition and familial tooth agenesis with colorectal-cancer predisposition, most often in a heterozygous carrier state.

Transcript
NM_004655.4
HGVS · transcript:coding
NM_004655.4:c.203G>A
GRCh38
chr17:65558418 C>T
GRCh37
chr17:63554536 C>T
VUS: BP1 (supporting) and BP4 (moderate) are the only met criteria, and together they fall short of the generic ACMG/AMP Likely Benign combination.
Classification rationale
BP1BP4 VUS
AXIN2 c.203G>A missense · exon 2

VUS: BP1 (supporting) - AXIN2 germline disease is truncating-driven, so this missense change does not fit the established loss-of-function mechanism. VUS: BP4 (moderate) - REVEL 0.176 for p.(Arg68Gln) sits at or below the 0.183 moderate benign threshold. VUS: no Benign, Likely Benign, Pathogenic, or Likely Pathogenic combination is met by one supporting plus one moderate benign criterion with no pathogenic evidence.

BP1 + BP4 → VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_004655.4 · variants mapped to exon structure
AXIN2 NM_004655.4
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 22 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
BP1 supporting review Benign
Met at supporting: AXIN2 disease is caused primarily by truncating loss-of-function variants, so this missense change qualifies for BP1 benign support.
pvs1_gene_context: lof_mechanism_supported=true for AXIN2 germline disease; literature queries included 'AXIN2 truncating variant germline' and 'AXIN2 inherited disease loss of function'.OncoKB gene curation: AXIN2 is a tumor suppressor; 'Truncating mutations in AXIN2 have been associated with colorectal cancer'.Gene summary: AXIN2 acts as a tumor suppressor in the Wnt/beta-catenin destruction complex, with mutations linked to colorectal cancer and to familial tooth agenesis with predisposition to colorectal cancer.
BP4 moderate Benign
Met (moderate): REVEL 0.176 for p.(Arg68Gln) is at or below the <= 0.183 moderate BP4 benign threshold.
Consequence class: missense (NP_004646.3:p.(Arg68Gln)) - BP4 taken from the REVEL missense path only, not from SpliceAI; the two paths are never combined and no prediction is double-counted.REVEL v1.3 local lookup (revel block): score 0.176 for chr17:g.65558418C>T (GRCh38), the same allele as NC_000017.10:g.63554536C>T.Thresholds applied: BP4 supporting requires REVEL <= 0.29; moderate <= 0.183; strong <= 0.016 (ClinGen SVI REVEL calibration, Pejaver et al. 2022, Am J Hum Genet, PMID:36413997). 0.176 meets supporting and moderate but not strong.
Assessed · not applied · 17 not met · 5 not assessed
Pathogenic
PS1 Not met: no report of the p.Arg68Gln amino-acid change established as pathogenic - the only record is ClinVar 127940, classified uncertain significance.
PS2 Not assessed: no proband or parental testing exists in any source, and the sole variant report (PMID:31285513) provides no parental genotype data.
PS3 Not met: no functional assay exists for AXIN2 p.Arg68Gln - the only paper naming the variant (PMID:31285513) reports it as a class-3 VUS with zero functional testing.
PS4 Not met: the only case observation is 1 carrier among 158 attenuated-polyposis patients, with no control group or odds ratio establishing enrichment.
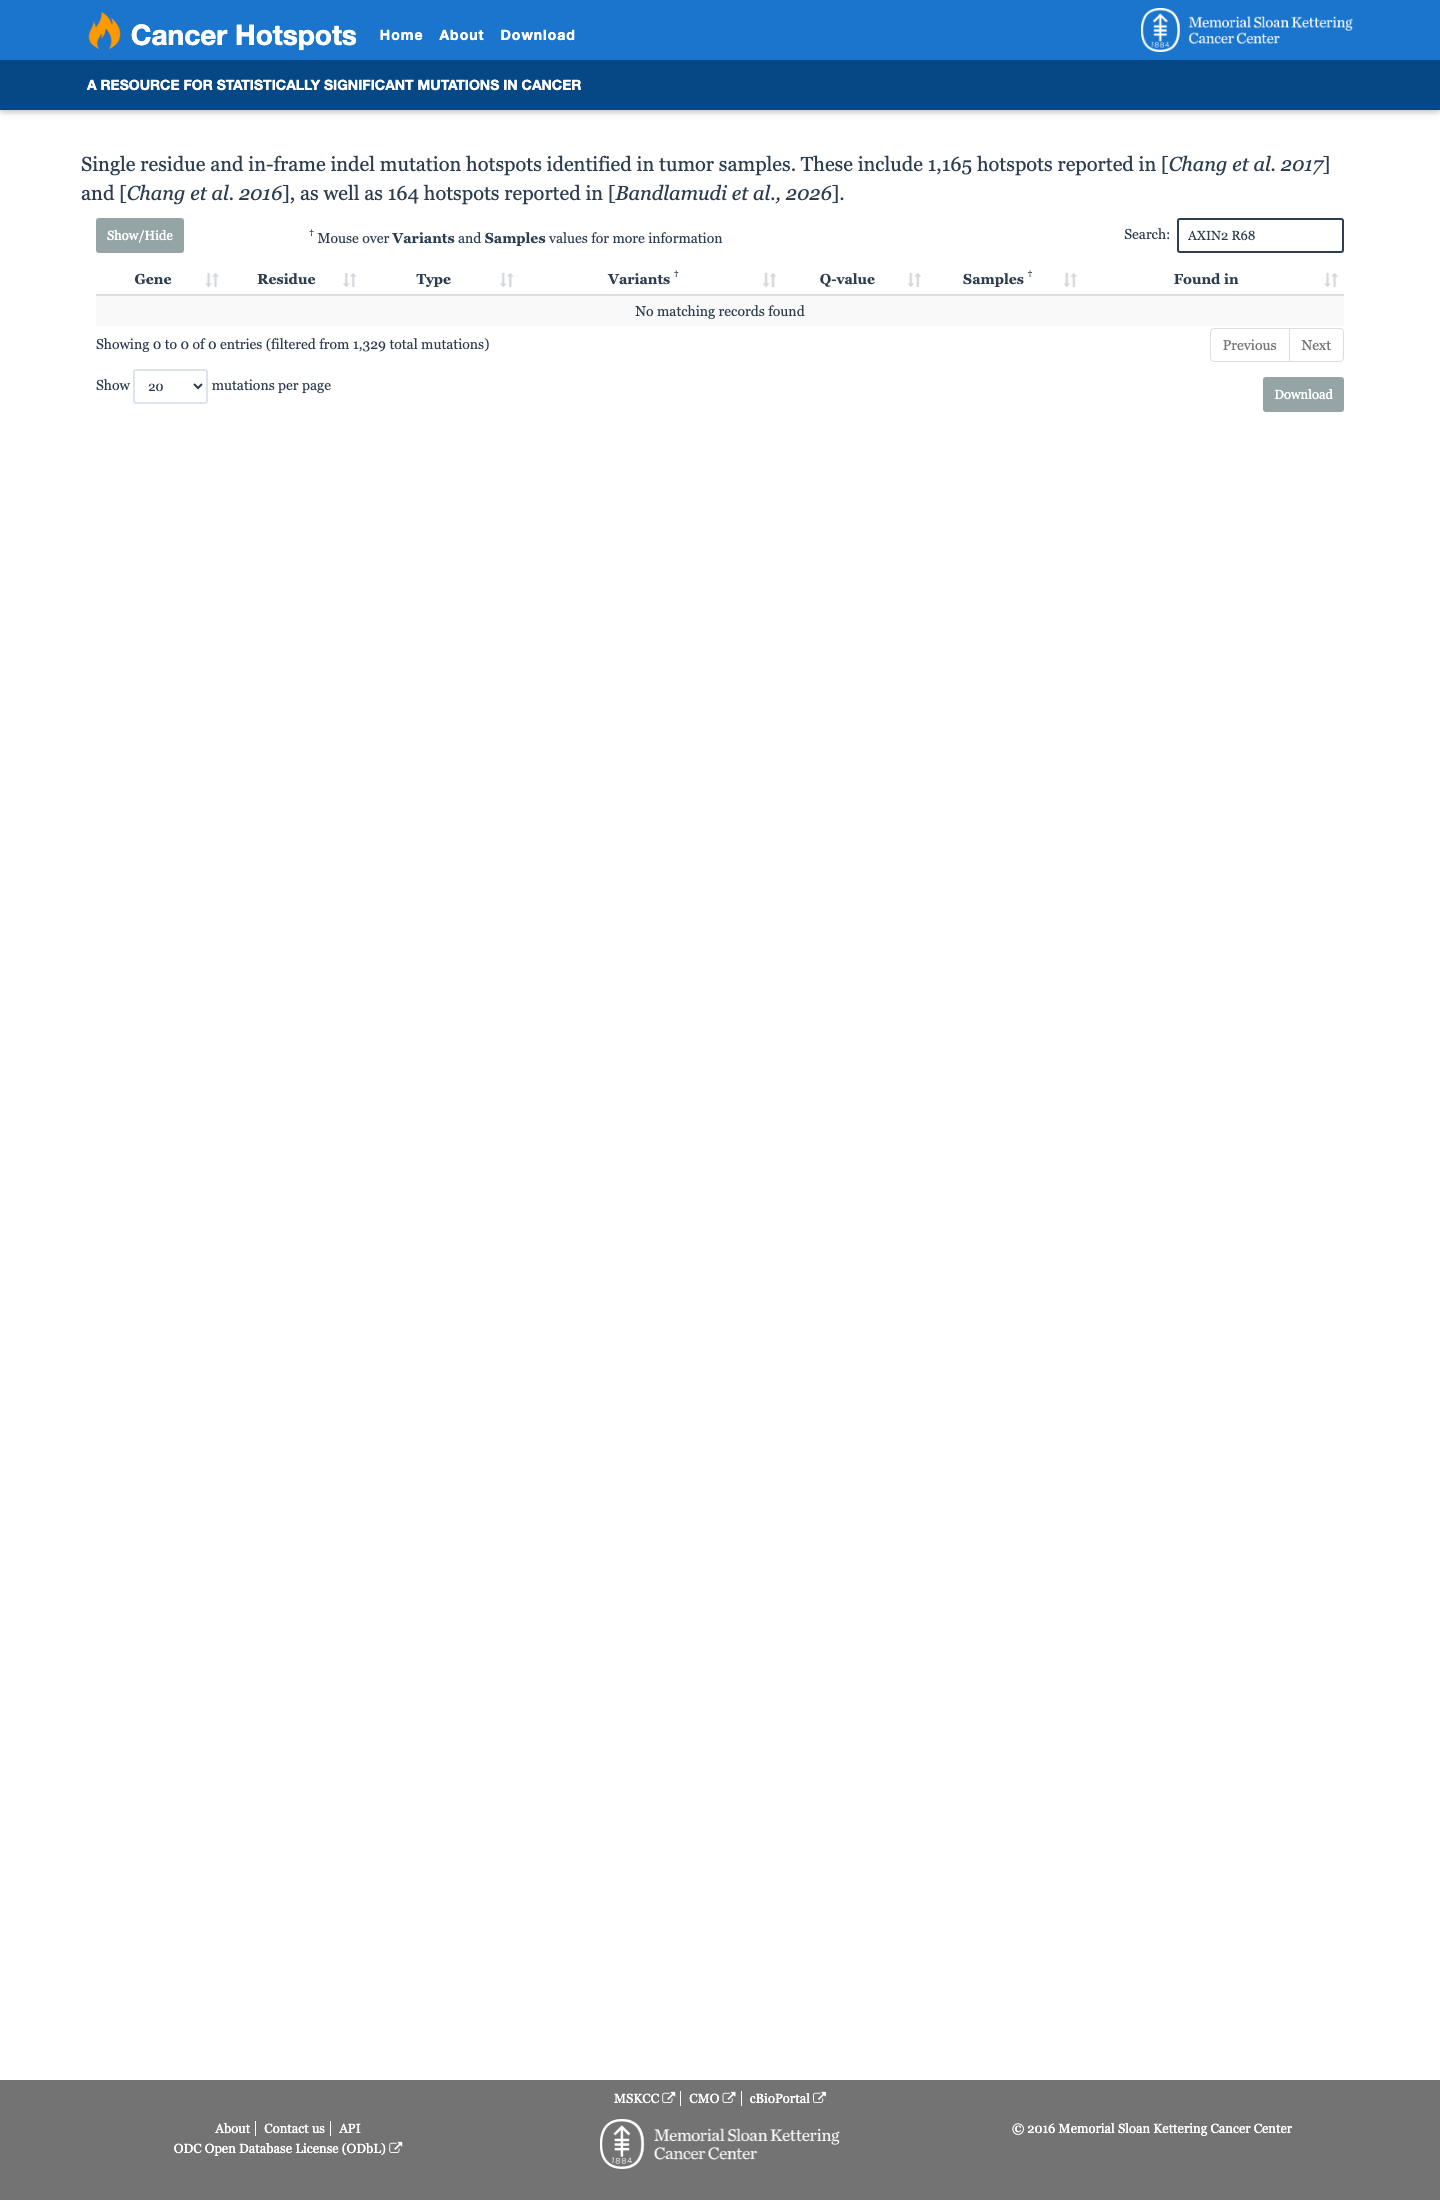
PM1 Not met: CancerHotspots is negative at AXIN2 R68 and the variant itself sits in gnomAD at 0.013% with no homozygotes, so no hotspot or variation-free critical domain is established.
PM2 Not met: gnomAD AF 0.000131 (v4.1) and 0.000135 (v2.1), corroborated by a published European frequency of 2e-04, exceed the <=0.0001 PM2 threshold.
PM3 Not met: no pathogenic AXIN2 allele in trans, and 0 homozygotes among 37/274,598 gnomAD v2.1 alleles.
PM5 Not met: no missense change other than p.Arg68Gln is reported at codon 68, so the required pathogenic same-residue comparator does not exist.
PM6 Not assessed: no parental testing or family data exist anywhere, and the sole variant report (PMID:31285513) documents no assumed de novo event.
PP1 Not assessed: no pedigree or genotyped affected relative exists in any source, so zero informative co-segregation meioses can be counted for c.203G>A.
PP2 Not met: AXIN2 disease is driven by truncating loss-of-function variants rather than missense, and no low benign-missense-rate constraint is established for the gene.
PP3 Not met: REVEL 0.176 for p.(Arg68Gln) sits far below the >= 0.644 supporting PP3 threshold.
PP4 Not assessed: no proband phenotype or family history is documented, and the only disease context (adenomatous polyposis) is genetically heterogeneous across APC, MUTYH and other genes.
PP5 Not met: ClinVar record 127940 has zero expert-panel submissions and no Pathogenic or Likely pathogenic assertion for this exact variant.
Benign
BA1 Not met: gnomAD v4.1 AF 0.000131 and v2.1 AF 0.000135 are roughly two orders of magnitude below the 5% BA1 threshold.
BS1 Not met: the highest gnomAD frequency, v4.1 grpmax FAF 0.000278, is ~36-fold below the 1% BS1 threshold.
BS2 Not met: zero homozygotes in gnomAD v2.1 (37 alleles) and v4.1 (210 alleles), and AXIN2 disease is adult-onset with incomplete penetrance.
BS3 Not met: no functional assay of AXIN2 p.Arg68Gln exists - the only protein-function statements are in-silico (ClinVar SCV000149781, SCV002584759), which cannot satisfy BS3.
BS4 Not assessed: no family or non-segregation data exist in any source, and submitters' likely-benign classifications alone cannot establish BS4.
BP2 Not met: no pathogenic AXIN2 variant reported in cis or in trans with c.203G>A; the single carrier was heterozygous for this variant alone.
BP5 Not met: no affected individual carrying this variant has a documented alternate molecular basis for disease in any reviewed source.
BP6 Not met: the two Likely benign ClinVar submissions for this variant are ordinary laboratory assertions, with zero expert-panel submissions on record.
N/A · 4 PVS1 · PM4 · BP3 · BP7
Research & evidence
Population frequency
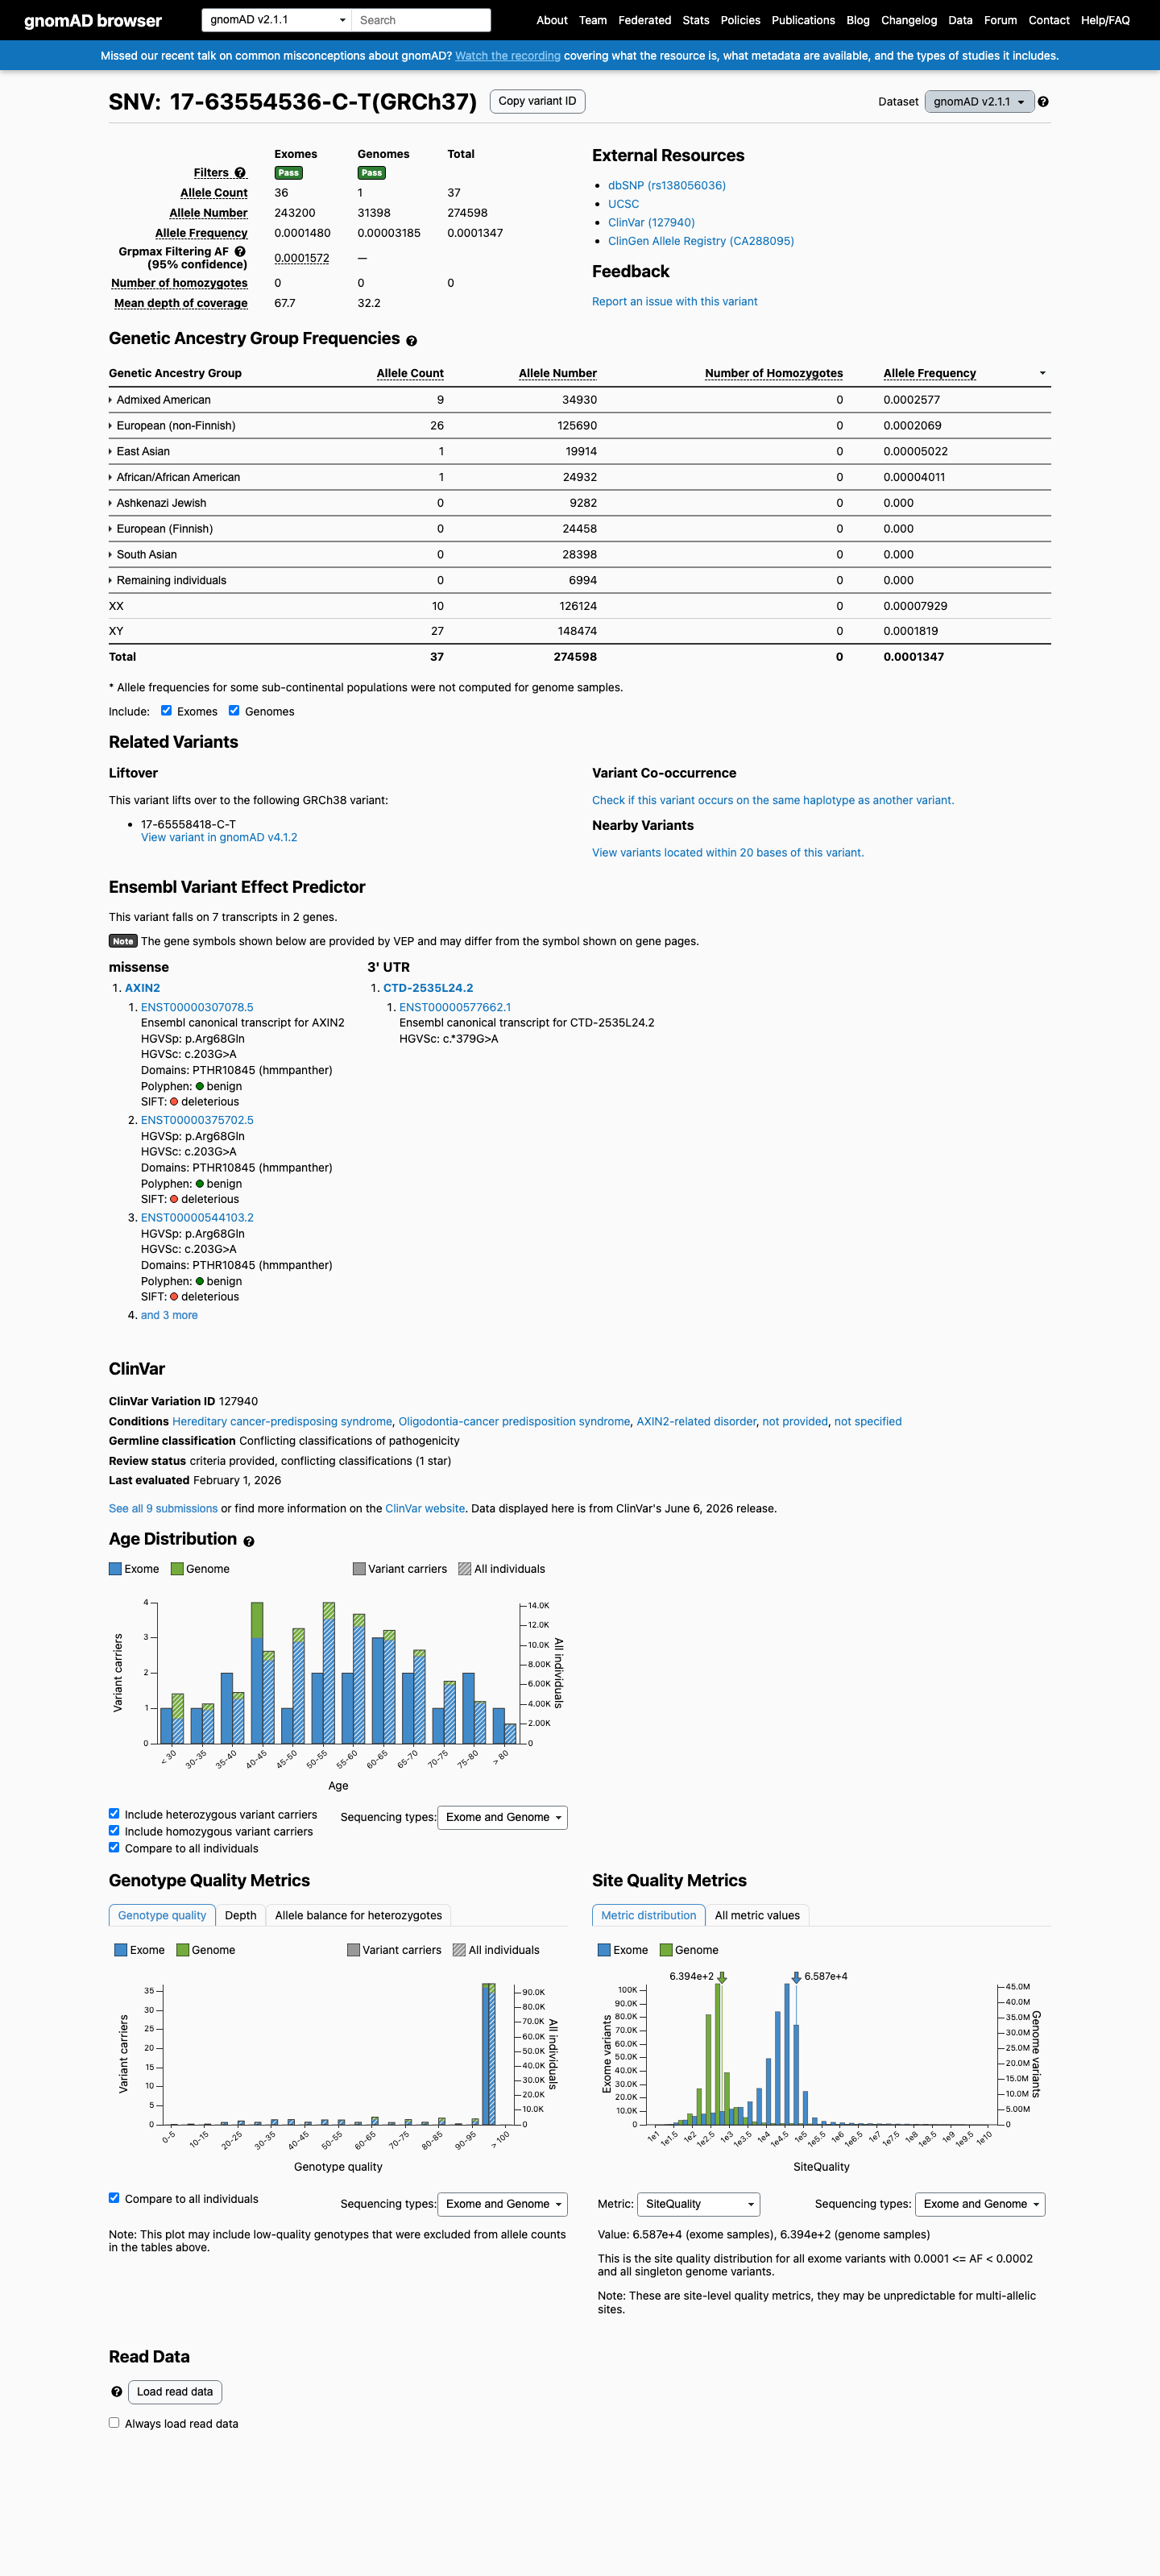
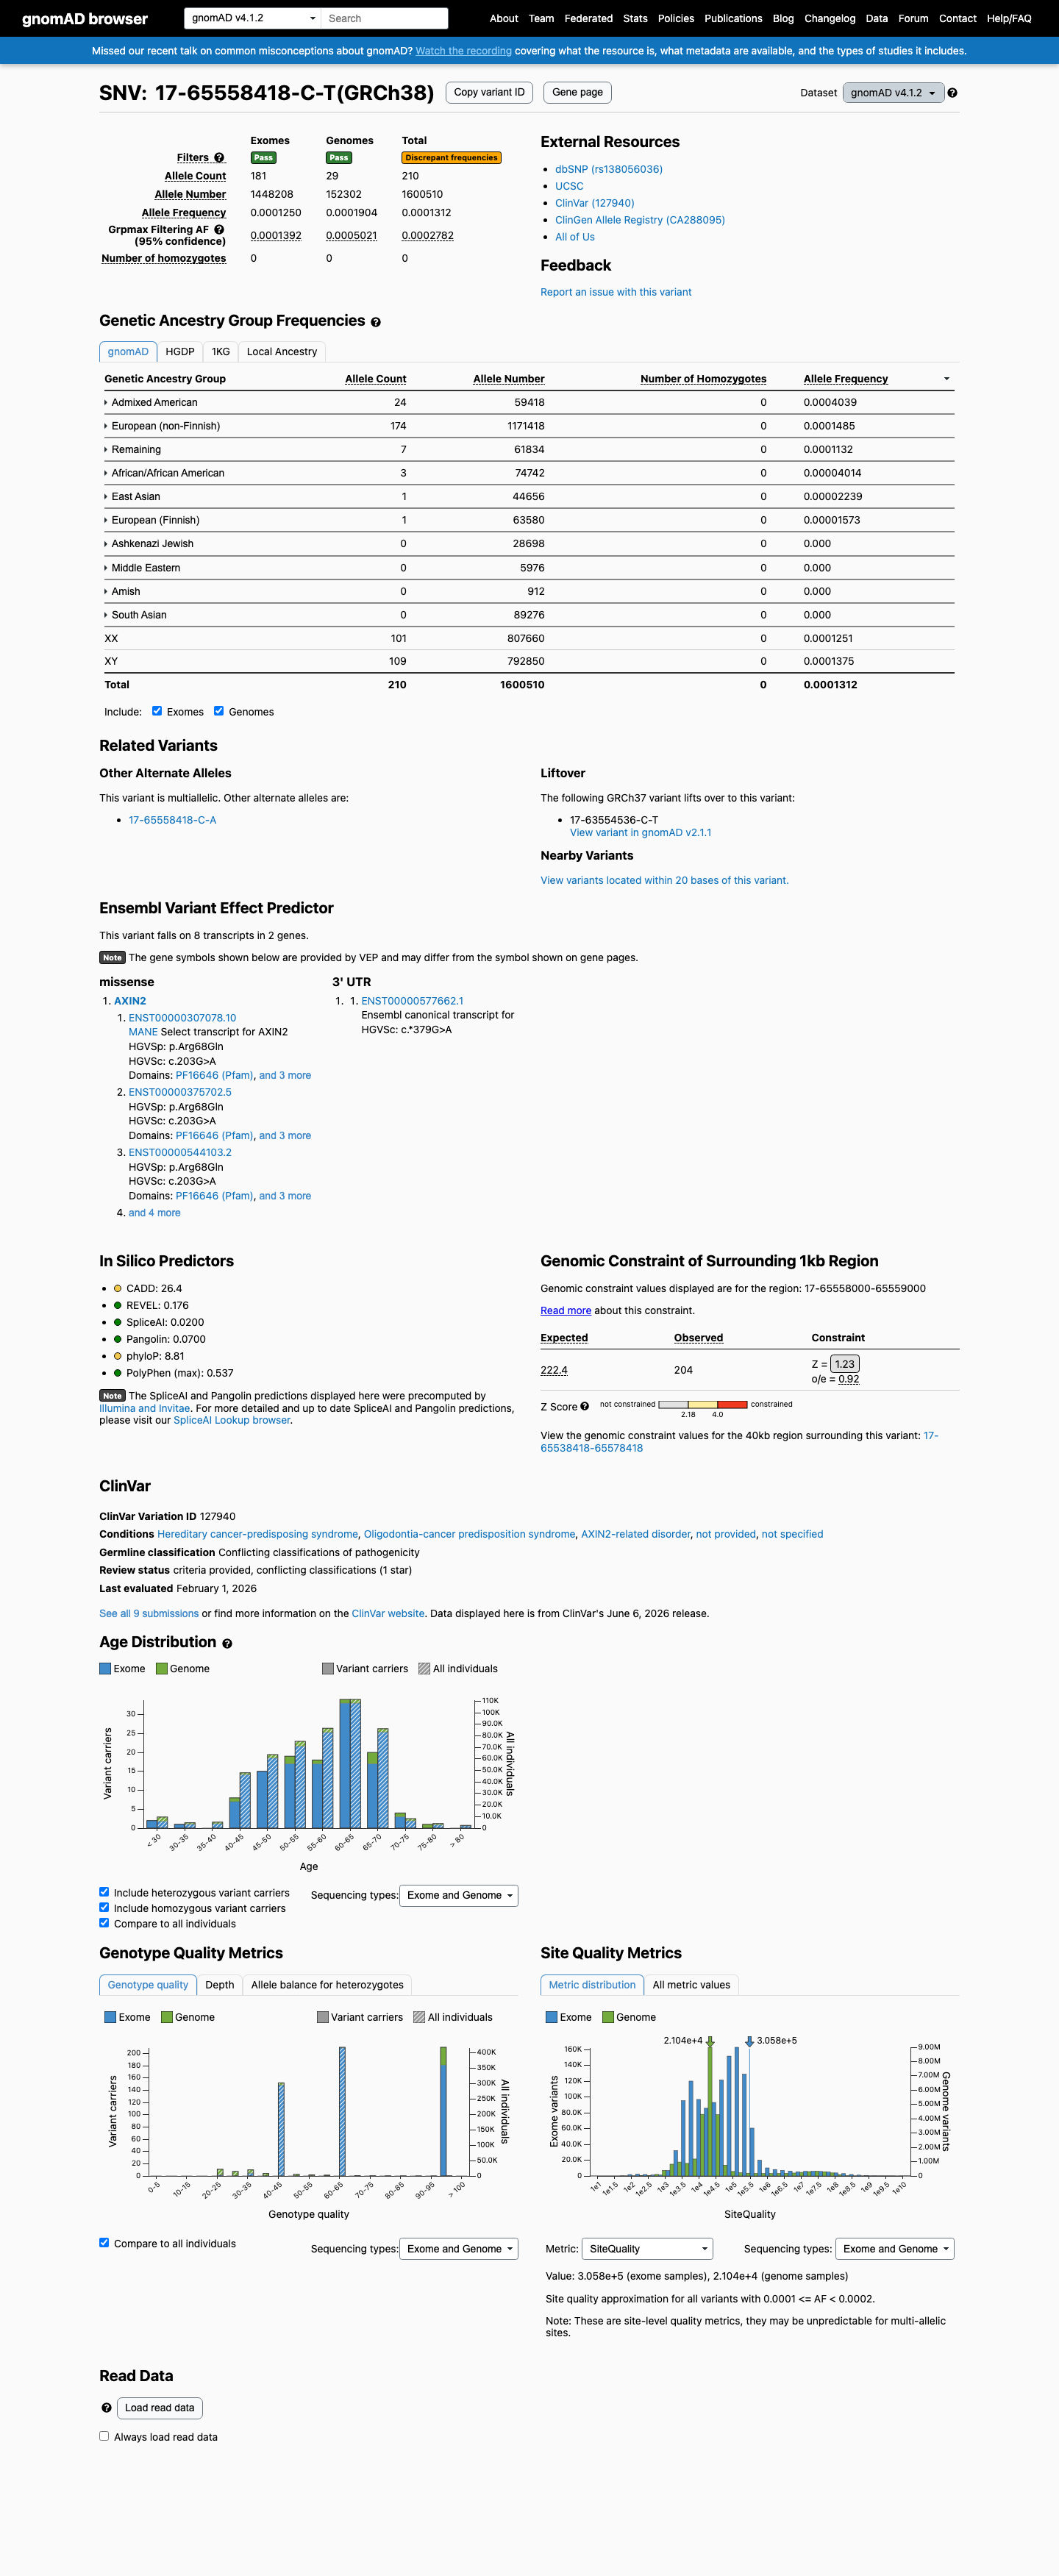
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 0.000131208; MAF= 0.01312%, 210/1600510 alleles, homozygotes = 0) and has highest observed frequency in the Admixed American population (AF= 0.000403918; MAF= 0.04039%, 24/59418 alleles, homozygotes = 0); grpmax FAF= 0.00027822.
v2.1
This variant is present in gnomAD v2.1 (AF= 0.000134742; MAF= 0.01347%, 37/274598 alleles, homozygotes = 0) and has highest observed frequency in the Admixed American population (AF= 0.000257658; MAF= 0.02577%, 9/34930 alleles, homozygotes = 0); grpmax FAF= 0.00015724.
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.013% · 210 / 1,600,510
0 hom · FAF 0.028%
Admixed American
24 / 59,418
0.04%
European (non-Finnish)
174 / 1,171,418
0.015%
Remaining individuals
7 / 61,834
0.011%
African/African American
3 / 74,742
0.004%
East Asian
1 / 44,656
0.0022%
European (Finnish)
1 / 63,580
0.0016%
+ 4 not observed (Amish, Middle Eastern, South Asian, Ashkenazi Jewish)
gnomAD v2.1
0.013% · 37 / 274,598
0 hom · FAF 0.016%
Admixed American
9 / 34,930
0.026%
European (non-Finnish)
26 / 125,690
0.021%
East Asian
1 / 19,914
0.005%
African/African American
1 / 24,932
0.004%
+ 4 not observed (Ashkenazi Jewish, European (Finnish), Remaining individuals, South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
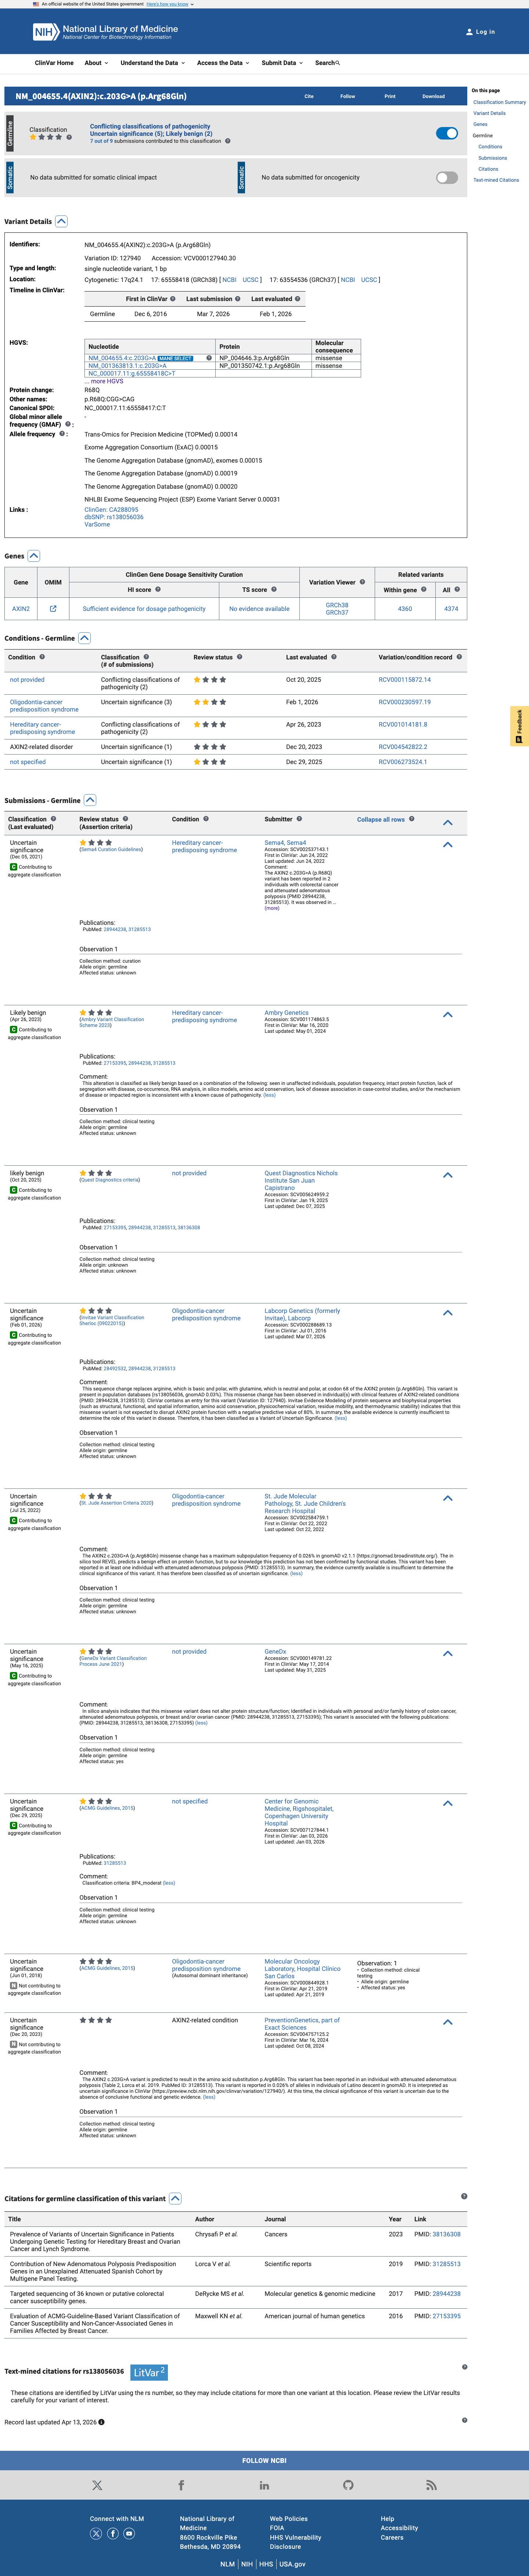
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Uncertain significance (6 clinical laboratories) and as Likely benign (1 clinical laboratory) and as likely benign (1 clinical laboratory). (ClinVarID = 127940)
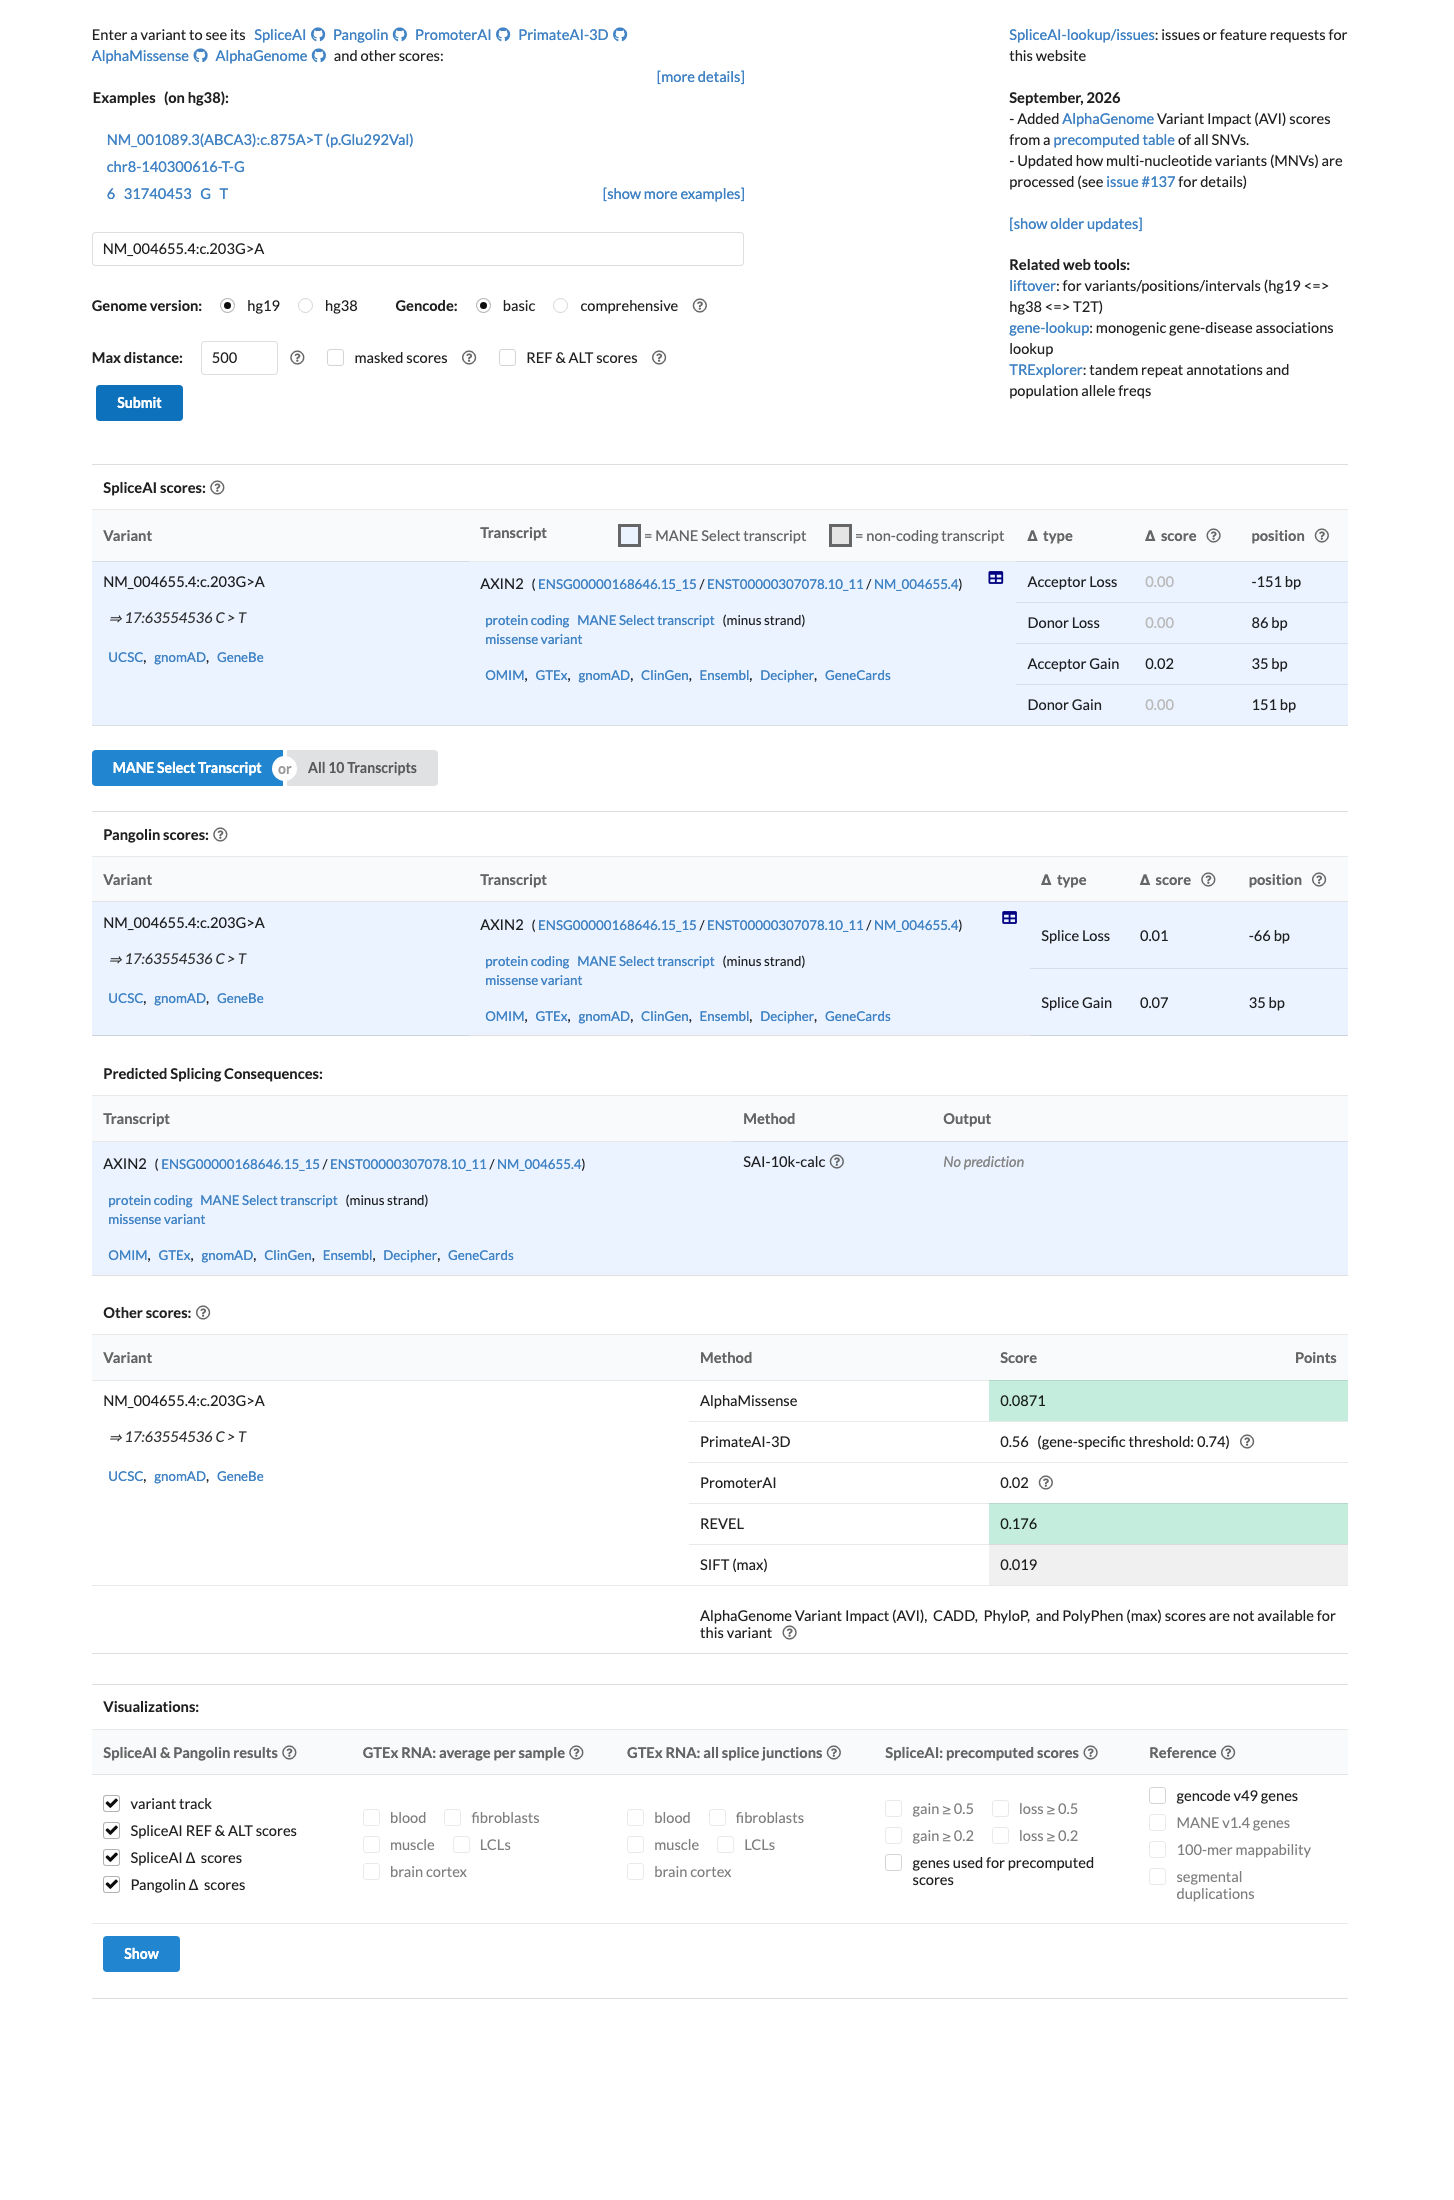
SpliceAI screenshot
In silico
SpliceAI returned NO scores for this variant, so no SpliceAI-based splice prediction is available. This is missing data, NOT evidence of absent splice impact: it must not be used to support BP4 or to argue against PP3/PVS1. Pangolin scores may be present but are not calibrated for PP3/BP4 here. REVEL score = 0.176. BayesDel score = -0.262489.
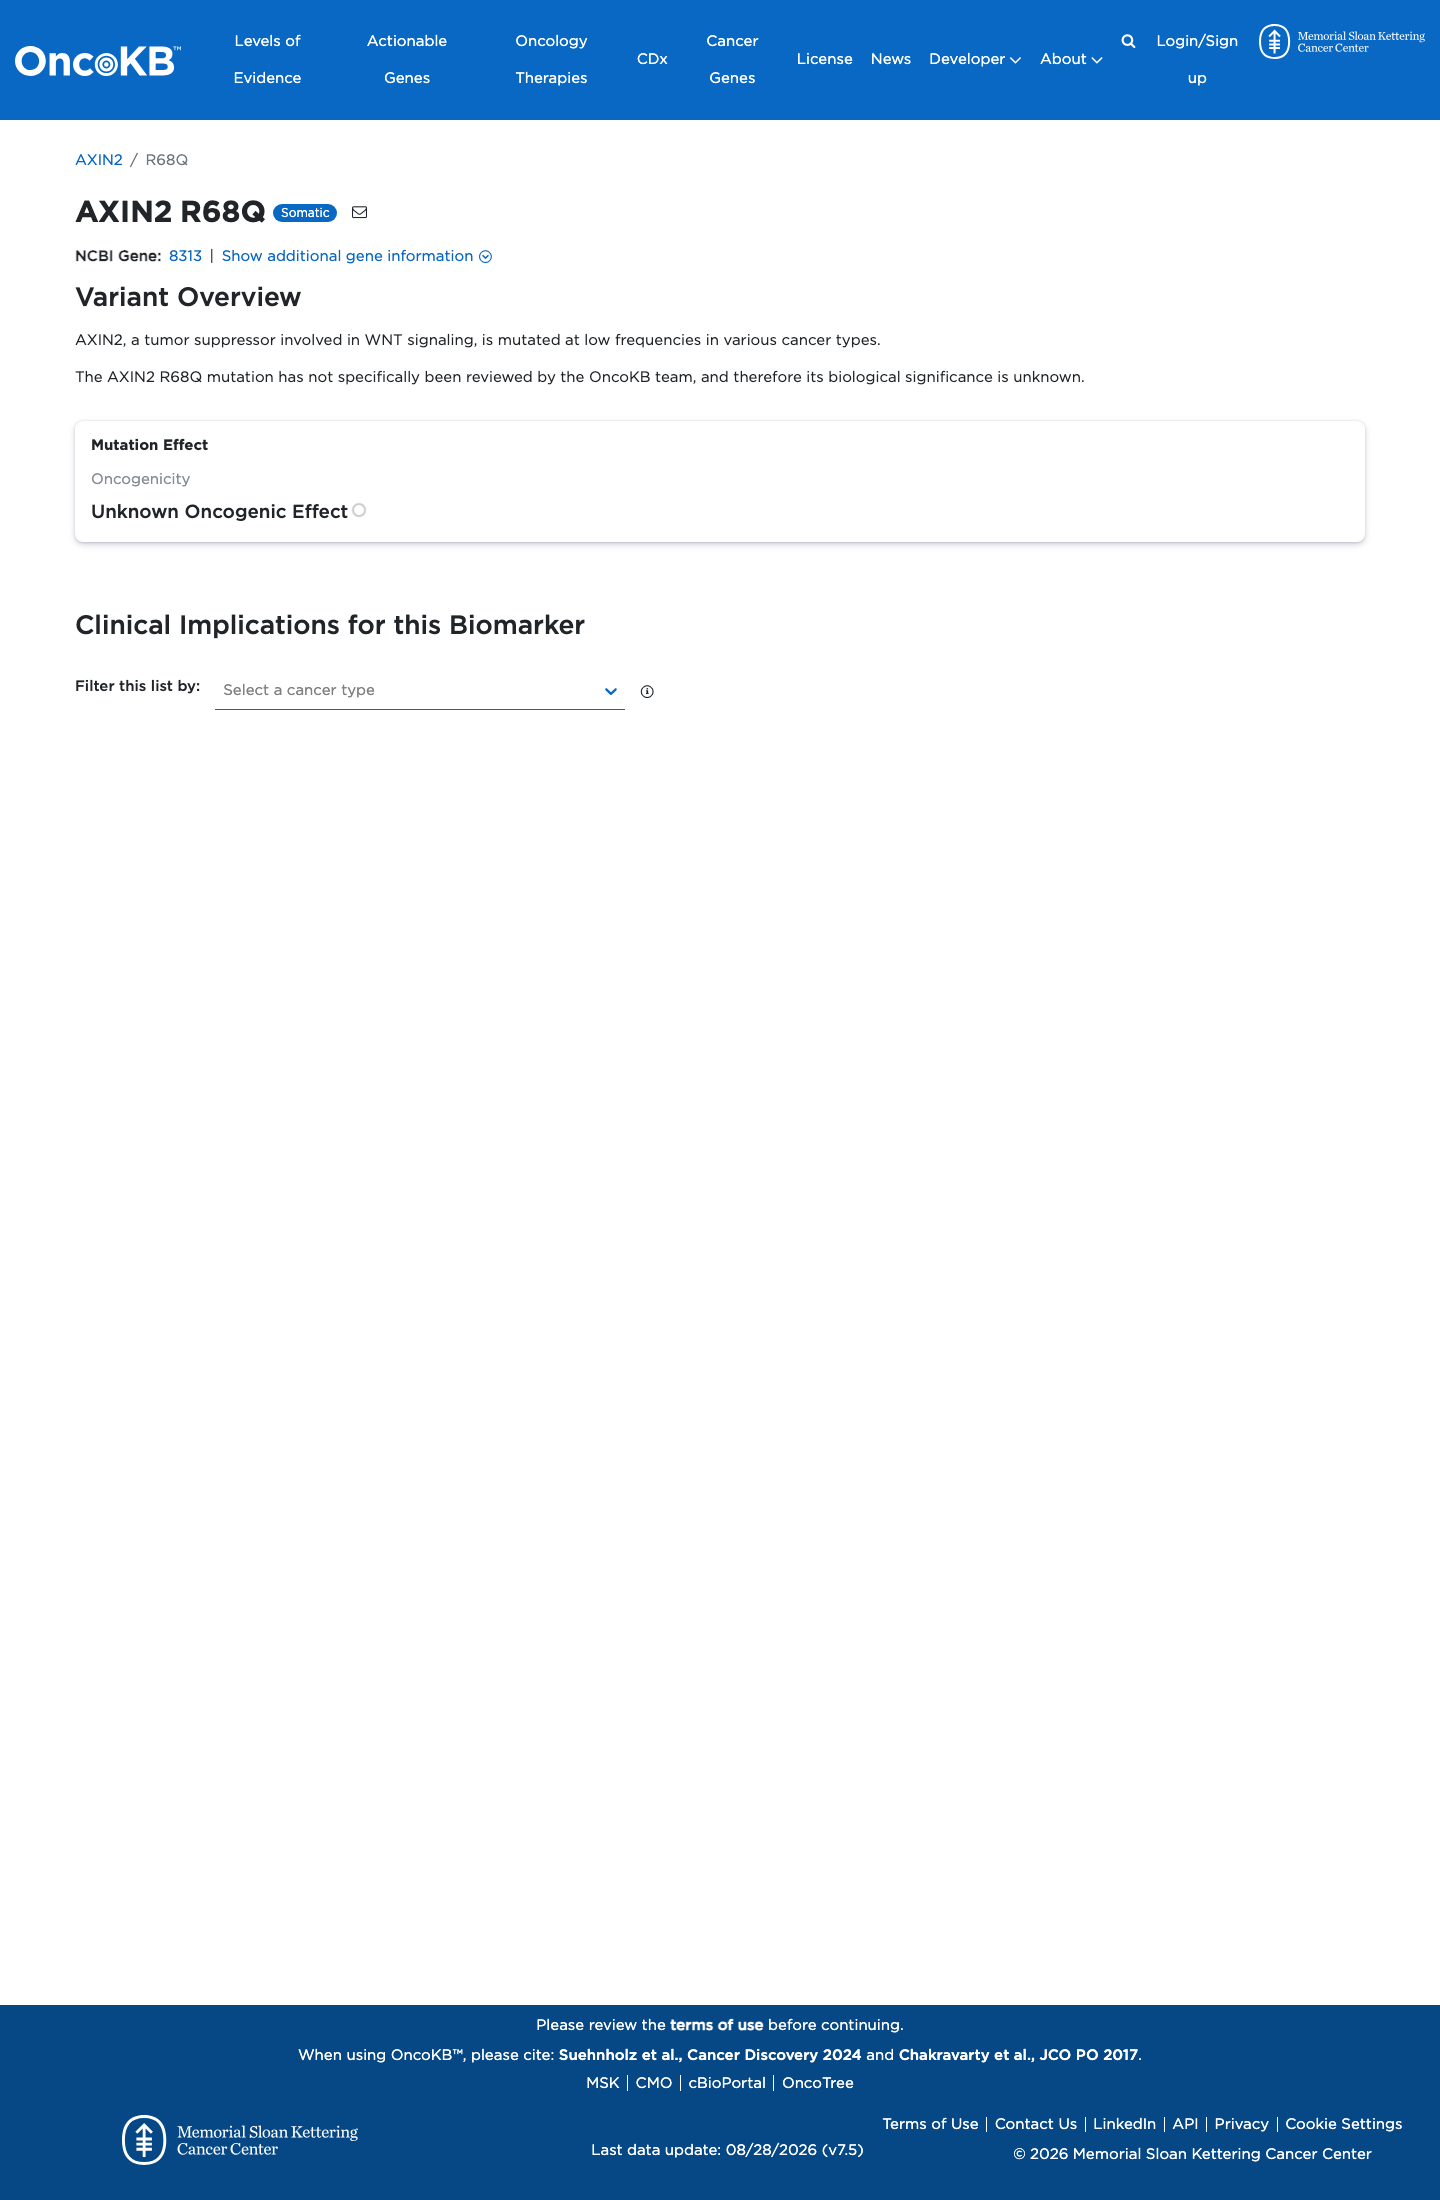
Functional / OncoKB screenshot
Functional Unknown Oncogenic Effect
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. AXIN2, a tumor suppressor involved in WNT signaling, is mutated at low frequencies in various cancer types.
OncoKB ↗
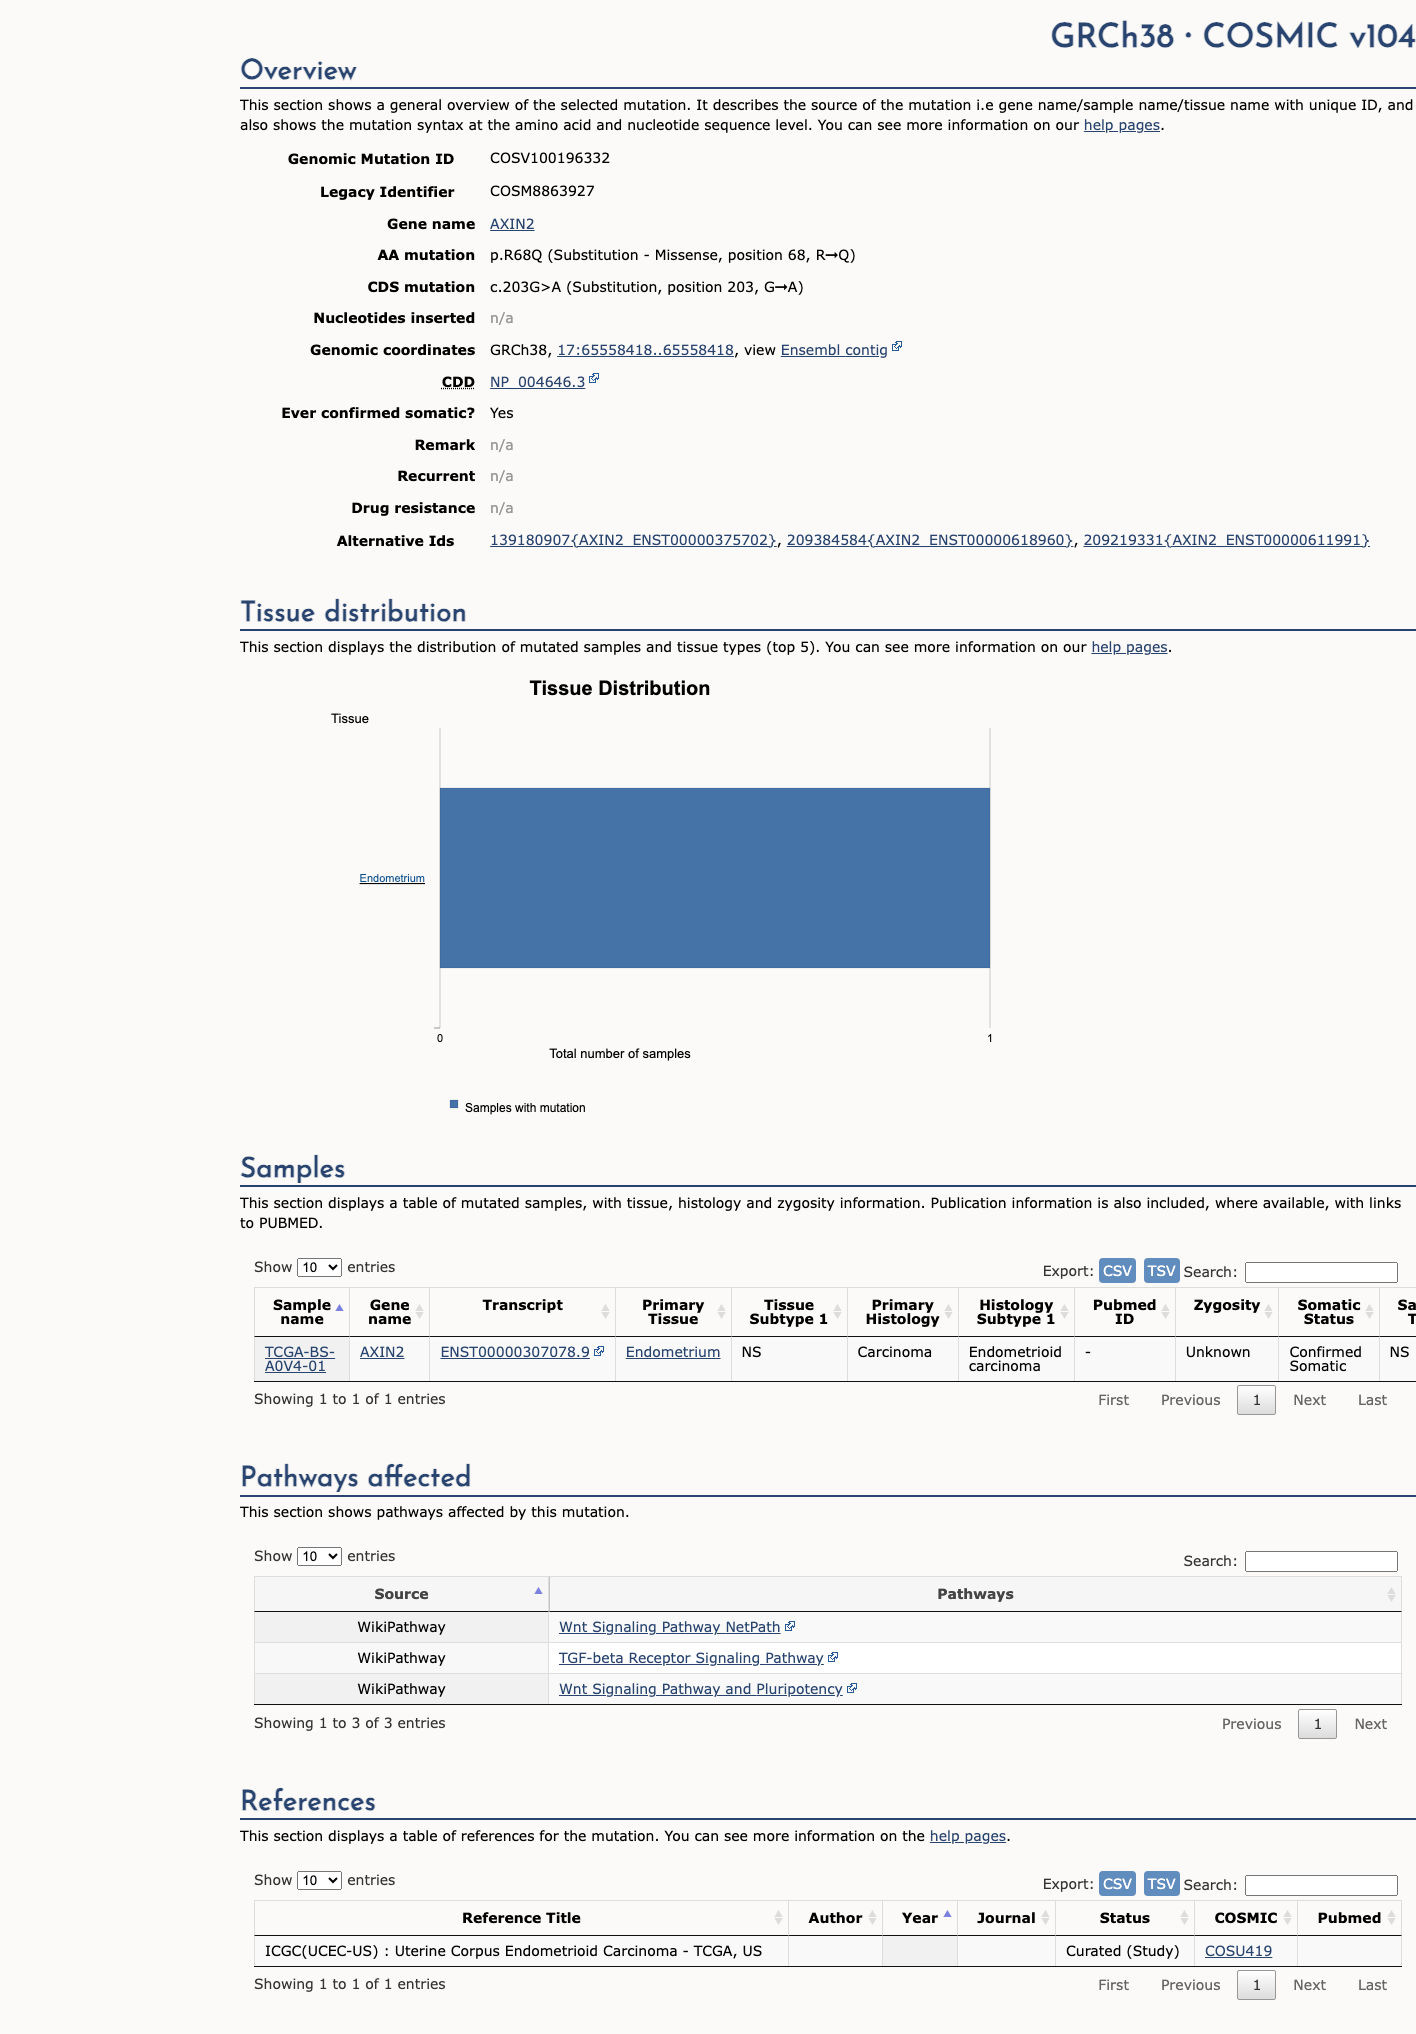
COSMIC screenshot
COSMIC
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has previously been reported in somatic cancers (COSMIC; COSV100196332, n = 1 times).
Hotspots
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
4papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 3 further PMIDs triaged but not cited — see Sources & references.
Contribution of New Adenomatous Polyposis Predisposition Genes in an Unexplained Attenuated Spanish Cohort by Multigene Panel Testing.
Searched
c.203G>Ac.203G > Ap.(Arg68Gln)p.(R68Q)rs138056036AXIN2 p.Arg68
Found
The AXIN2 variant c.203G>A p.(Arg68Gln) (rs138056036) was detected in one attenuated adenomatous polyposis patient (ID 79) in a Spanish cohort of 158 AAP subjects screened by a multigene NGS panel. It was heterozygous, had a gnomAD (European Non-Finnish) frequency of 2e-04, was annotated to the SMART protein domain AXIN1_TNKS, and was classified as class 3, i.e. a variant of uncertain significance under ACMG-SHERLOC criteria. It was not among the actionable findings; the paper states that none of the patients showed actionable (class-4/class-5) mutations in AXIN2. No functional, segregation, or somatic data specific to this variant are presented.
Variant
✓ Names this variant — characterised directly
Applied to
→BP4 moderate
79 c.203G > A p.(Arg68Gln) rs138056036 2e-04 AXIN1_TNKS 3
Location Table 2 (validated variants and classification according to ACMG-SHERLOC criteria), AXIN2 rows, patient ID 79; corroborated in Results (germline DNA screening) and Discussion ('None of the patients showed actionable mutations (class-4 and -5) in any of the genes associated with new polyposis syndromes (MSH3, NTHL1, POLD1, POLE or AXIN2)')  ·  Context NGS Haloplex custom panel (Agilent) sequencing of coding regions and intron-exon boundaries of APC, MUTYH, POLE, POLD1, NTHL1, MSH3 and AXIN2 in 158 unrelated Spanish attenuated adenomatous polyposis (AAP) patients (10-100 adenomas); all validated variants heterozygous; classification by ACMG-SHERLOC criteria with gnomAD and 1000 Genomes frequency data.  ·  full text
Rule & framework references · cited for criterion definitions, not variant evidence
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.
26467025 ↗ A Standardized DNA Variant Scoring System for Pathogenicity Assessments in Mendelian Disorders.
28944238 ↗ Targeted sequencing of 36 known or putative colorectal cancer susceptibility genes.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 3 PMIDs not cited in assessment
27153395 ↗ Evaluation of ACMG-Guideline-Based Variant Classification of Cancer Susceptibility and Non-Cancer-Associated Genes in Families Affected by Breast Cancer. CLINVAR
25394175 ↗ A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment. CLINVAR
38136308 ↗ Prevalence of Variants of Uncertain Significance in Patients Undergoing Genetic Testing for Hereditary Breast and Ovarian Cancer and Lynch Syndrome. CLINVAR