Back
NM_002439.5:c.1522A>G
p.Lys508Glu · MSH3
ACMG/AMP
0%
complete
Final classification
Likely Benign
BP1BP4
MSH3
c.1522A>G
p.Lys508Glu
missense · exon 10

MSH3 encodes a protein that partners with MSH2 to form MutS beta, a key component of the DNA mismatch repair system that recognizes and corrects errors made during DNA replication. It acts as a tumor suppressor: loss of MSH3 function leads to mismatch repair deficiency, an elevated mutation rate, and increased cancer risk, and the gene is frequently altered in mismatch repair-deficient colorectal cancers. Defects in MSH3 are linked to susceptibility to endometrial cancer, and inherited mutations are associated with an autosomal recessive form of familial adenomatous polyposis, with minimal evidence for a role in Lynch syndrome. Tumors with mismatch repair deficiency, including those involving MSH3, tend to respond well to immune checkpoint inhibitor therapies.

This variant

MSH3 encodes the MutS-beta (MSH2-MSH3) mismatch repair protein, and MSH3-related disease is an autosomal recessive, loss-of-function adenomatous polyposis phenotype in which truncating rather than missense alleles are established as causal, so this missense p.(Lys508Glu) change is not of the class expected to cause MSH3-related disease.

Transcript
NM_002439.5
HGVS · transcript:coding
NM_002439.5:c.1522A>G
GRCh38
chr5:80728919 A>G
GRCh37
chr5:80024738 A>G
Likely Benign: BP1 (supporting, missense in a truncating-spectrum gene) plus BP4 (supporting, REVEL 0.191) reach two supporting benign criteria under generic ACMG/AMP 2015.
Classification rationale
BP1BP4 Likely Benign
MSH3 c.1522A>G missense · exon 10

Likely Benign: BP1 (supporting) - p.(Lys508Glu) is a missense change in MSH3, whose established pathogenic variants are truncating (frameshift, nonsense, splice) against 1170 missense variants of uncertain significance. Likely Benign: BP4 (supporting) - REVEL 0.191 sits in the benign range (<=0.29) for this missense variant.

BP1 + BP4 → Likely Benign
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_002439.5 · variants mapped to exon structure
MSH3 NM_002439.5
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 19 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
BP1 supporting Benign
Met at supporting: MSH3 disease is caused by truncating variants - ClinVar MSH3 P/LP records are frameshift/nonsense/splice versus 1170 missense VUS.
ClinVar MSH3 pathogenic/likely pathogenic records are truncating: e.g. c.1790dup (p.Ser598fs), c.1660dup (p.Met554fs), c.1808C>G (p.Ser603Ter), c.1651C>T (p.Gln551Ter), c.910-1G>A, c.793-1G>A - frameshift, nonsense and canonical splice variants only in the established pathogenic set.ClinVar MSH3 missense records (1200-record sample): 1170 Uncertain significance, 28 conflicting, only 2 singleton P/LP assertions - missense change is not an established disease mechanism for MSH3.The variant under assessment is a missense substitution, NM_002439.5:c.1522A>G, NP_002430.3:p.(Lys508Glu).
BP4 supporting Benign
Met at supporting strength: REVEL 0.191 is within the <=0.29 BP4 supporting threshold but above the <=0.183 moderate cut-off.
Path selection: missense consequence (NP_002430.3:p.(Lys508Glu)) selects the REVEL missense path for BP4; SpliceAI max delta 0.002 was not used as BP4 support.Local REVEL v1.3 lookup returned revel_score 0.191 for chrom 5, pos 80728919 (GRCh38), ref A, alt G; found=true.REVEL thresholds applied verbatim from the ClinGen SVI calibration (Pejaver et al. 2022, Am J Hum Genet, PMID:36413997): BP4 supporting <=0.29, moderate <=0.183, strong <=0.016.
Assessed · not applied · 13 not met · 6 not assessed
Pathogenic
PS1 Not met: p.Lys508Glu is not established as pathogenic - ClinVar shows 5 Likely benign and 1 VUS submission and no pathogenic assertion.
PS2 Not assessed: no parental-testing data exist, so the confirmed de novo PS2 requires (PMID:25741868) is undeterminable; gnomAD v2.1 shows 106 alleles and v4.1 two homozygotes.
PS3 Not met: no assay has tested p.(Lys508Glu), and OncoKB reports no variant-specific functional evidence for this change.
PS4 Not met: no case-control enrichment exists, as the variant is absent from the 1231-case CRC cohort and present in gnomAD v4.1 at 392/1,612,578 alleles.
PM1 Not met: residue 508 lies in MSH3 MutS domain II but is no hotspot and is not free of benign variation (gnomAD popmax AF 0.44%, 2 homozygotes).
PM2 Not met: total allele frequency 0.024-0.038% in gnomAD v4.1/v2.1 exceeds the 0.01% PM2 rarity threshold, with the African/African American popmax at 0.44%.
PM3 Not met: no pathogenic variant was observed in trans with MSH3 c.1522A>G, and no phase-resolved proband data supports an in-trans configuration.
PM5 Not met: the only other codon-508 missense variant, p.Lys508Arg, has conflicting ClinVar classifications and is not established pathogenic.
PM6 Not assessed: no parental testing or de novo observation is documented, so PM6's assumed-de-novo premise (PMID:25741868) cannot be evaluated; the variant recurs in gnomAD (106 v2.1 alleles).
PP1 Not assessed: no pedigree or relative genotypes exist and no paper reports this variant in a family, so PP1 co-segregation (PMID:25741868) cannot be evaluated.
PP2 Not met: MSH3 shows no missense constraint (gnomAD oe_mis 1.08, mis_z -1.29) and its established pathogenic variants are truncating, not missense.
PP3 Not met: REVEL 0.191 for this missense variant is below the >=0.644 PP3 supporting threshold.
PP4 Not assessed: no proband phenotype or family history was recorded for this case, so PP4's requirement for a highly specific single-gene disease phenotype cannot be tested.
Benign
BA1 Not met: the highest frequency at usable sample size is 0.44% (gnomAD v4.1 African/African American), roughly tenfold below the 5% stand-alone benign threshold.
BS1 Not met: the highest frequency at usable sample size is 0.44% (gnomAD v4.1 African/African American), about 2.3-fold below the generic 1% BS1 threshold.
BS3 Not met: no functional assay of p.(Lys508Glu) exists, so preserved MSH3 function has never been experimentally demonstrated.
BS4 Not assessed: no pedigree or relative genotypes exist to demonstrate non-segregation, and absence of family data is not BS4 evidence (PMID:25741868).
BP2 Not met: neither a cis nor a trans pathogenic variant was observed with MSH3 c.1522A>G, and no phase-resolved partner allele is reported anywhere.
BP5 Not assessed: no case carrying this variant was reported anywhere in the evidence, so the alternate-molecular-basis observation BP5 requires could not be evaluated.
N/A · 7 PVS1 · PM4 · PP5 · BS2 · BP3 · BP6 · BP7
Research & evidence
Population frequency · supports benign
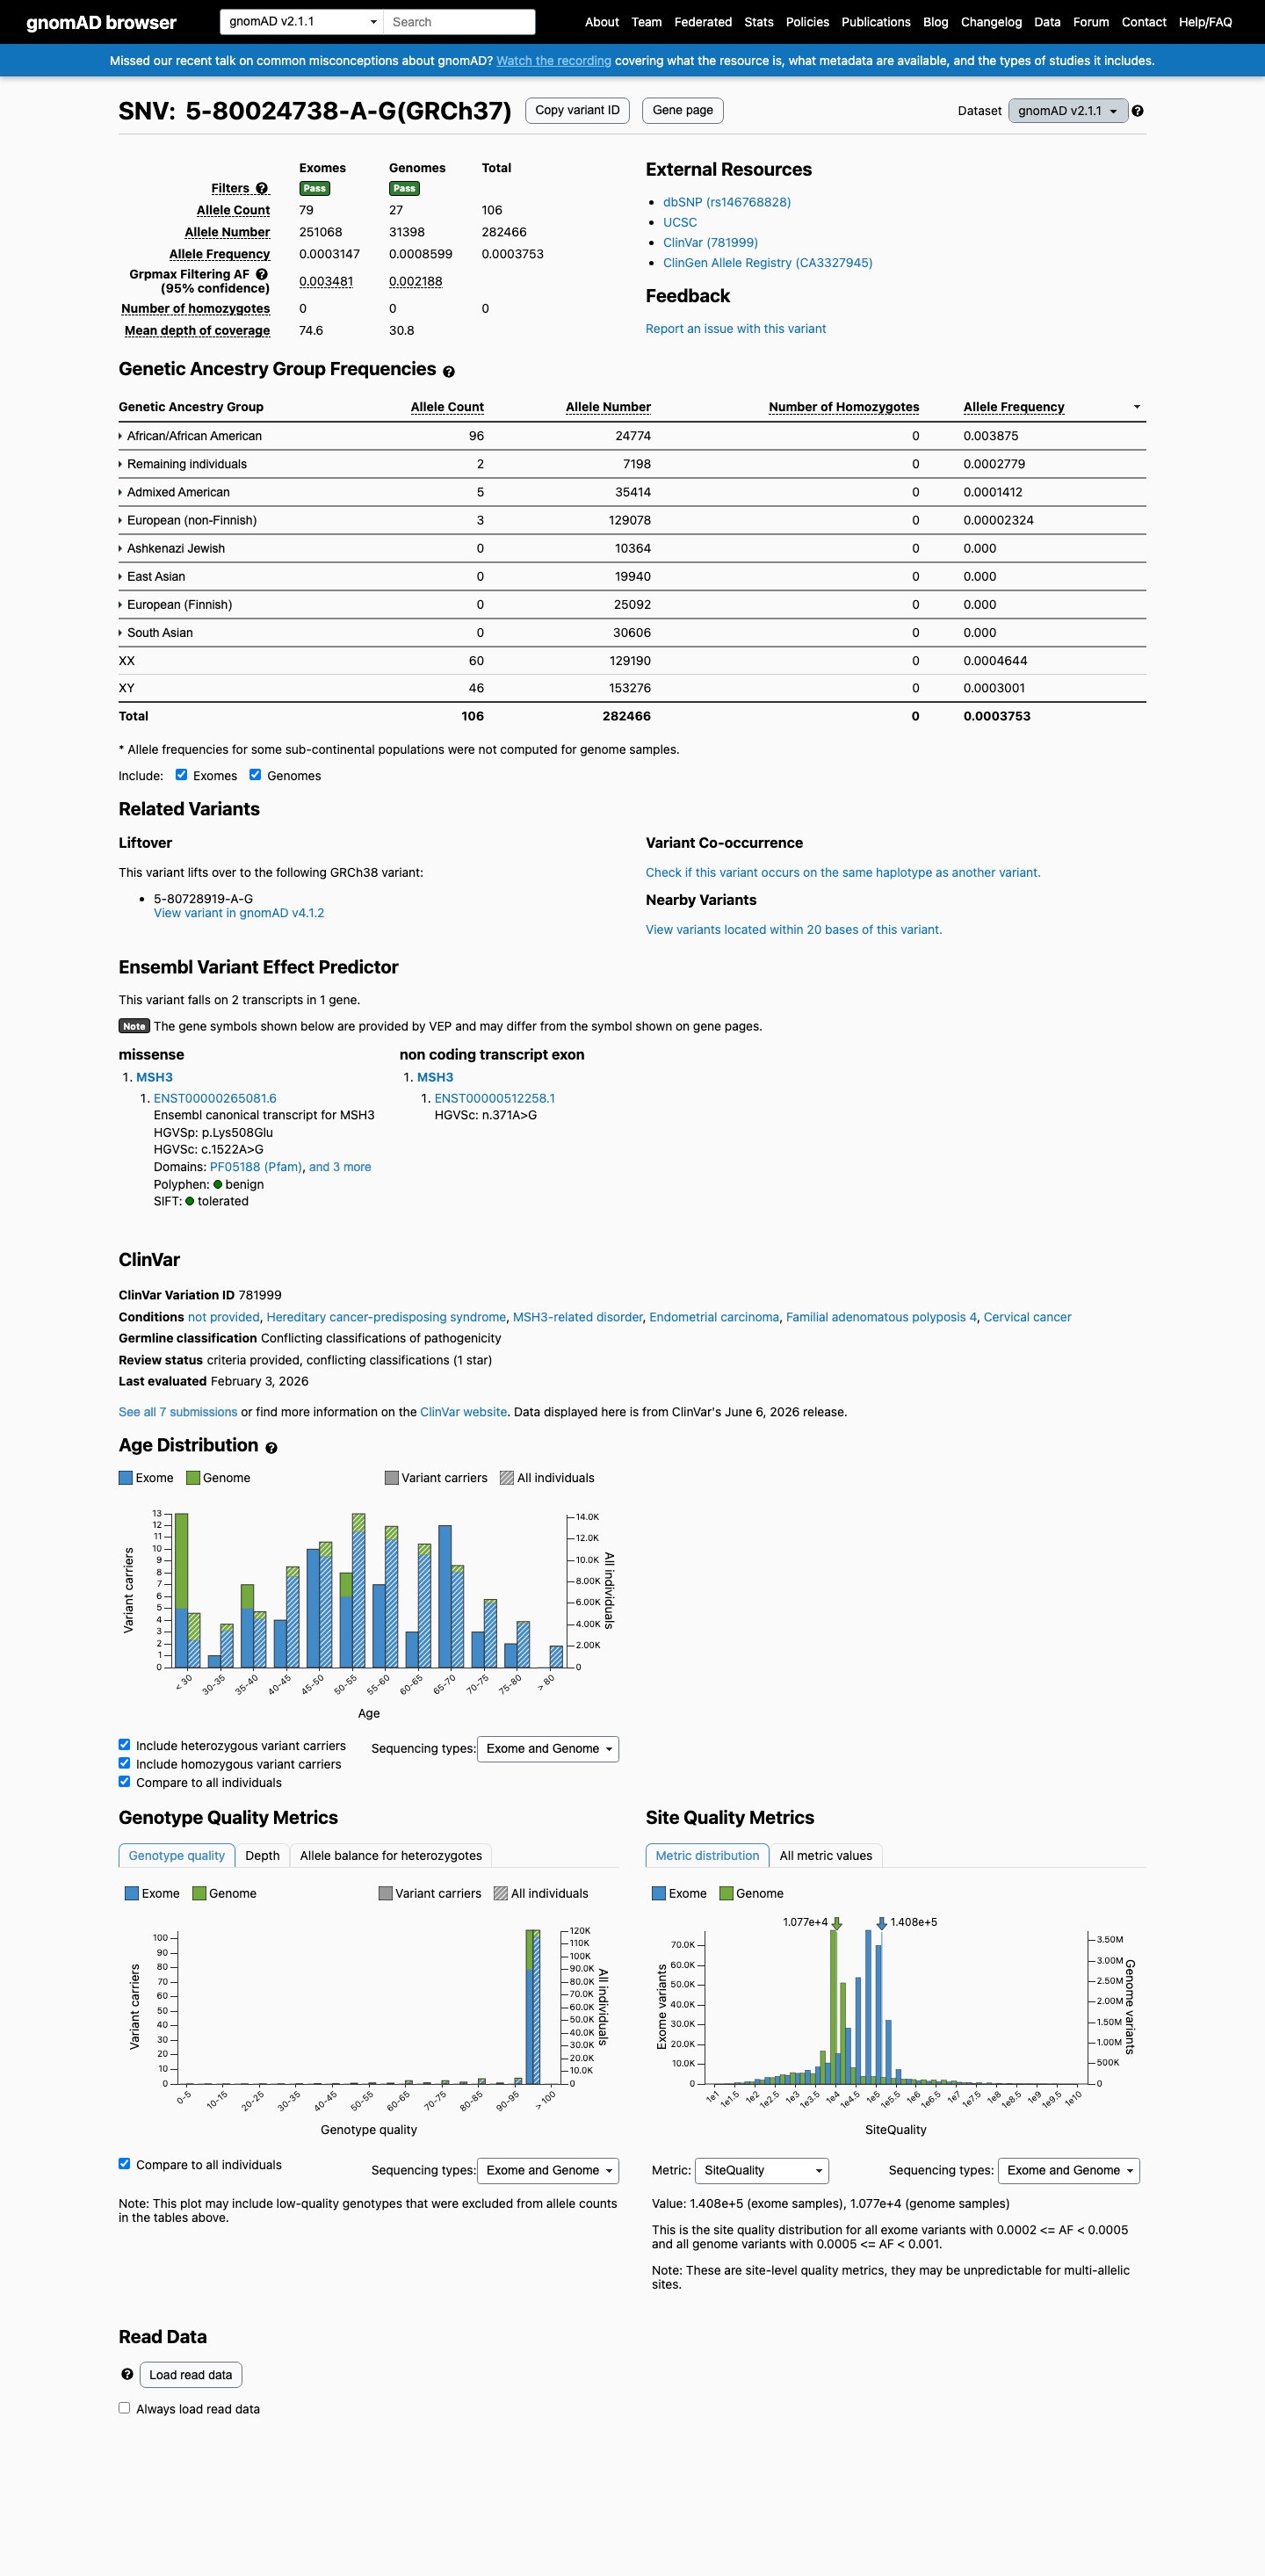
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
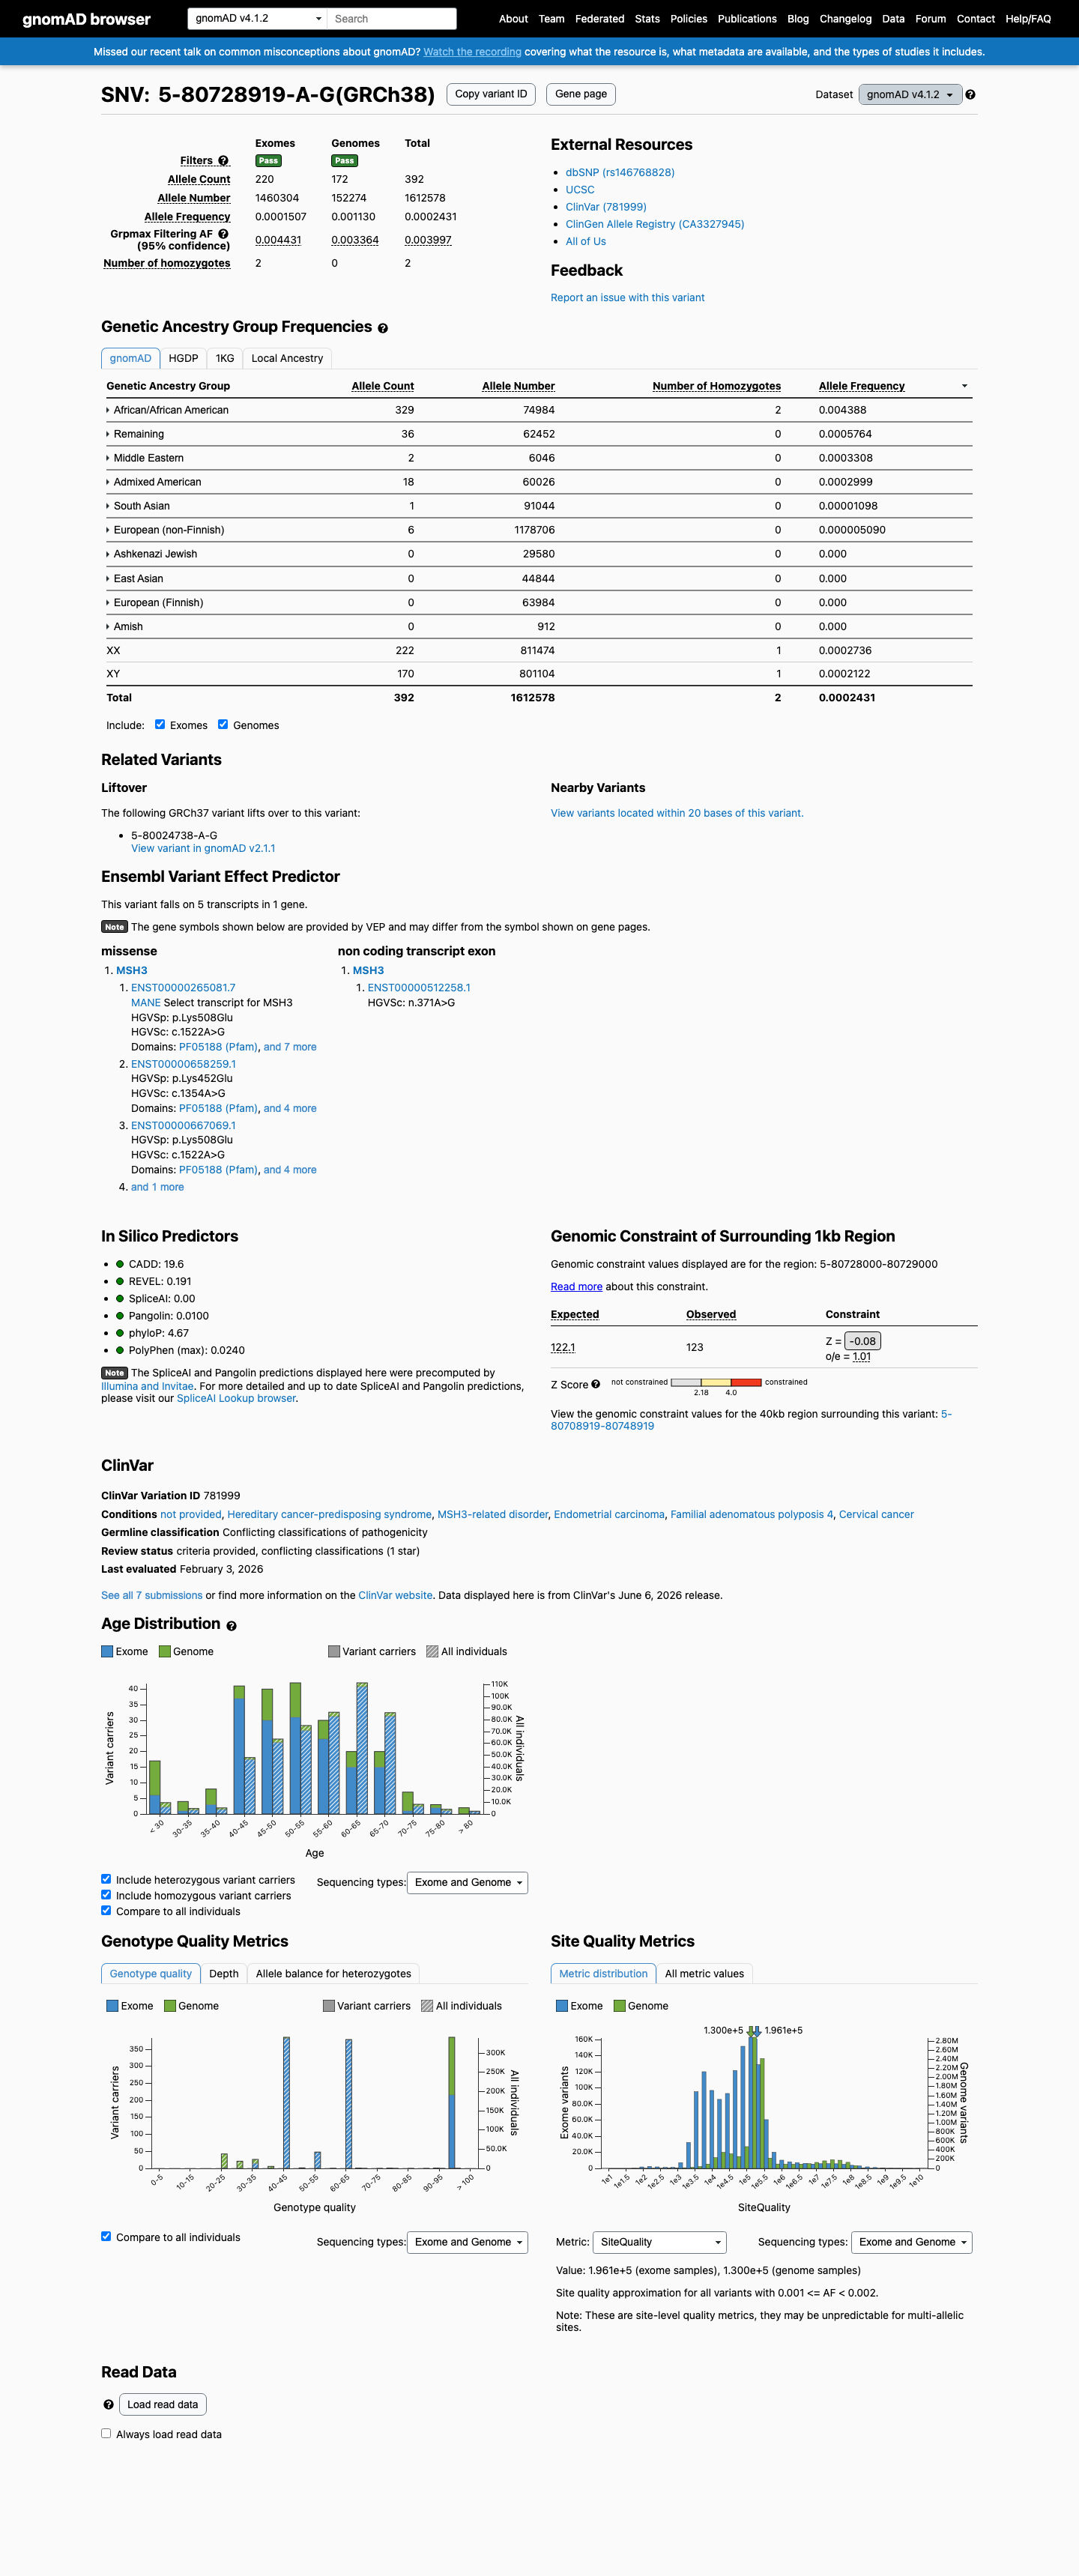
This variant is present in gnomAD v4.1 (AF= 0.000243089; MAF= 0.02431%, 392/1612578 alleles, homozygotes = 2) and has highest observed frequency in the African/African American population (AF= 0.0043876; MAF= 0.43876%, 329/74984 alleles, homozygotes = 2); grpmax FAF= 0.00399731.
v2.1
This variant is present in gnomAD v2.1 (AF= 0.000375266; MAF= 0.03753%, 106/282466 alleles, homozygotes = 0) and has highest observed frequency in the African/African American population (AF= 0.00387503; MAF= 0.38750%, 96/24774 alleles, homozygotes = 0); grpmax FAF= 0.00348064.
🇨🇦 CA
This variant is present in gnomAD-Canada v1.0 (AF= 0.00016286644951140066, 3/18420 alleles, homozygotes = 0).
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.024% · 392 / 1,612,578
2 hom · FAF 0.4%
African/African American
329 / 74,984
0.44%
2 hom
Remaining individuals
36 / 62,452
0.058%
Middle Eastern
2 / 6,046
0.033%
Admixed American
18 / 60,026
0.03%
South Asian
1 / 91,044
0.0011%
European (non-Finnish)
6 / 1,178,706
0.00051%
+ 4 not observed (European (Finnish), Amish, East Asian, Ashkenazi Jewish)
gnomAD v2.1
0.038% · 106 / 282,466
0 hom · FAF 0.35%
African/African American
96 / 24,774
0.39%
Remaining individuals
2 / 7,198
0.028%
Admixed American
5 / 35,414
0.014%
European (non-Finnish)
3 / 129,078
0.0023%
+ 4 not observed (Ashkenazi Jewish, East Asian, European (Finnish), South Asian)
gnomAD Canada 🇨🇦
0.016% · 3 / 18,420
0 hom · FAF 0.08%
African/African American
3 / 1,020
0.29%
+ 8 not observed (Latino/Admixed American, Ashkenazi Jewish, East Asian, European (Finnish), Middle Eastern, European (non-Finnish), Remaining individuals, South Asian)
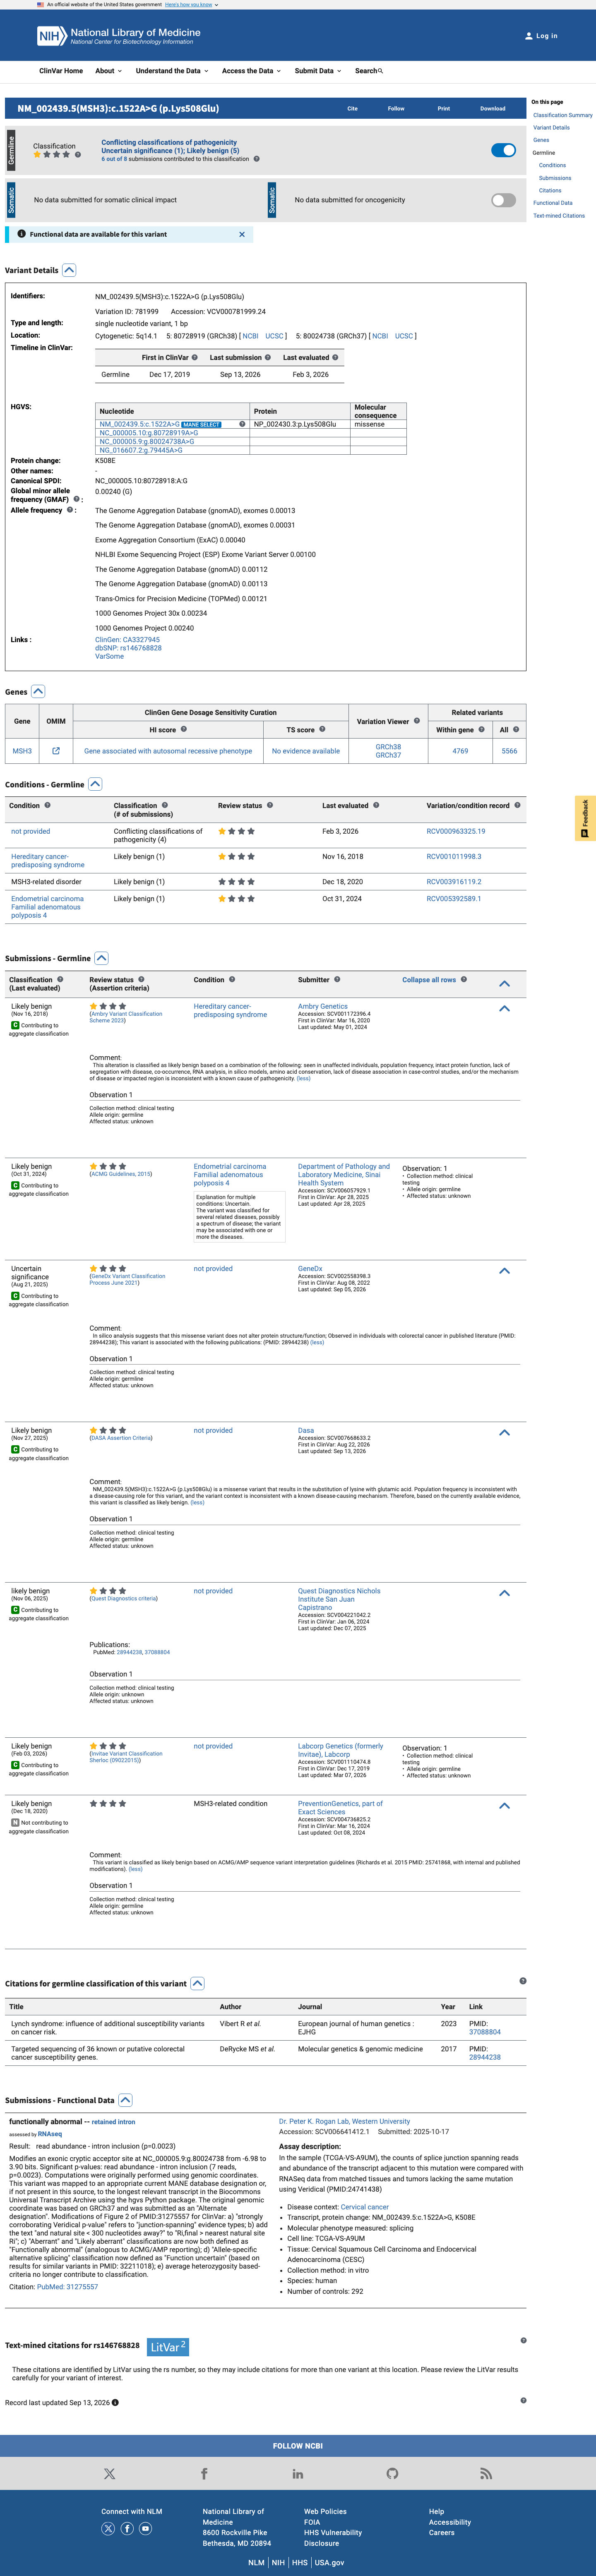
ClinVar screenshot
ClinVar
This variant is present in ClinVar (Variation ID: 781999); submission details unavailable.
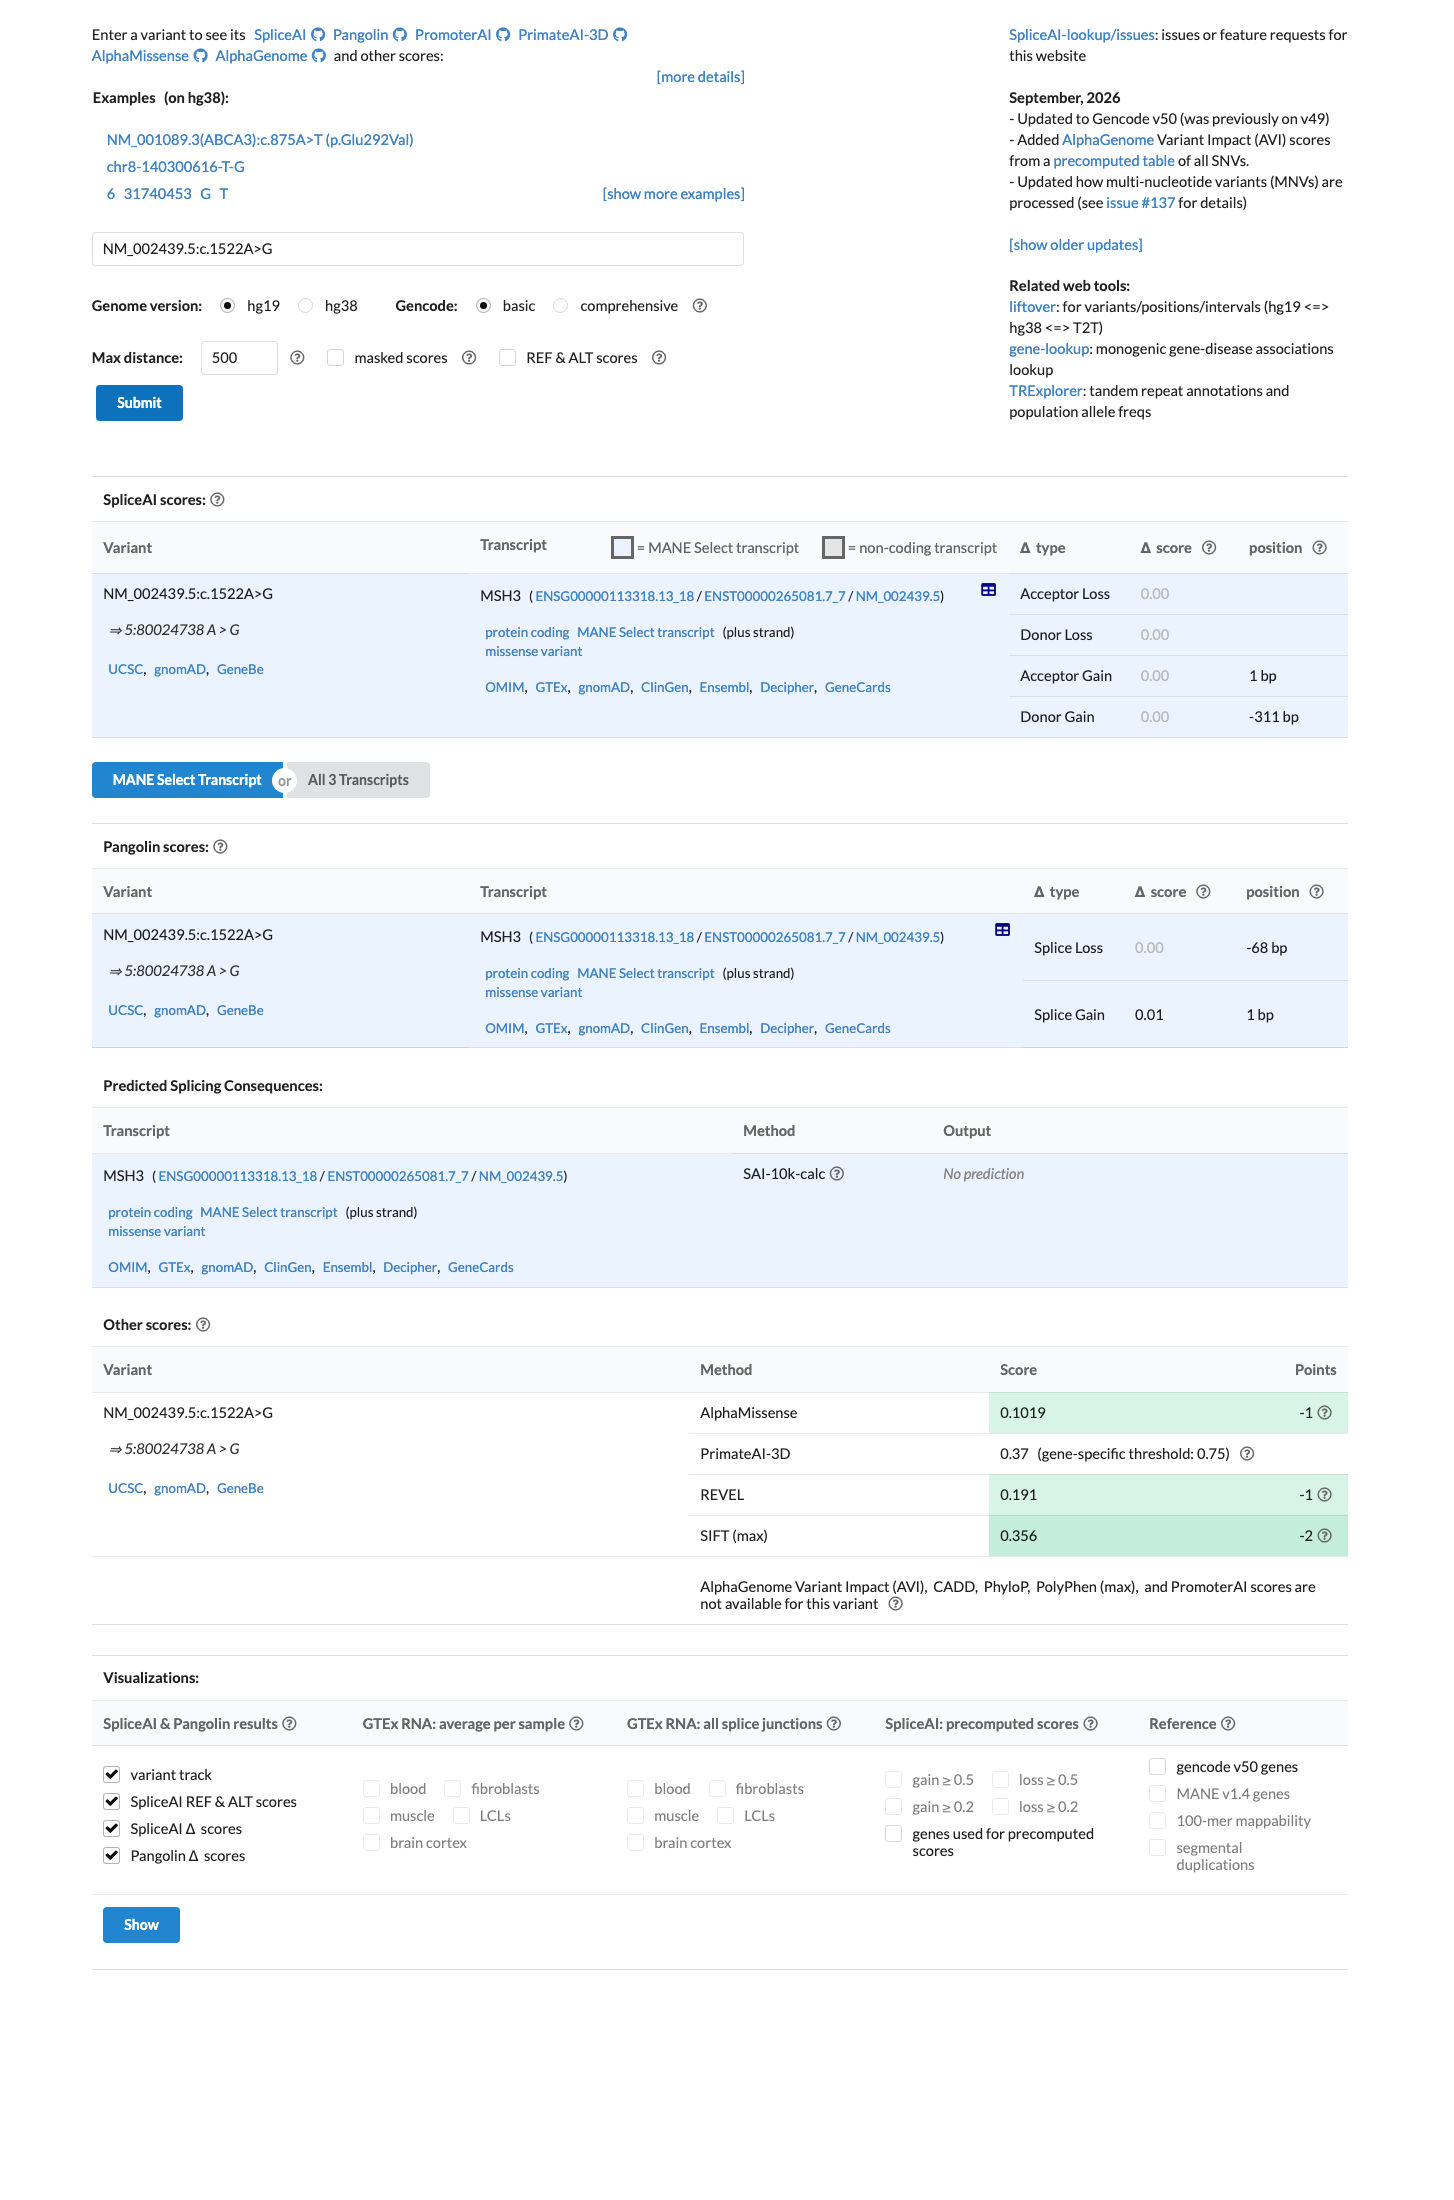
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.00). REVEL score = 0.191. BayesDel score = -0.230708.
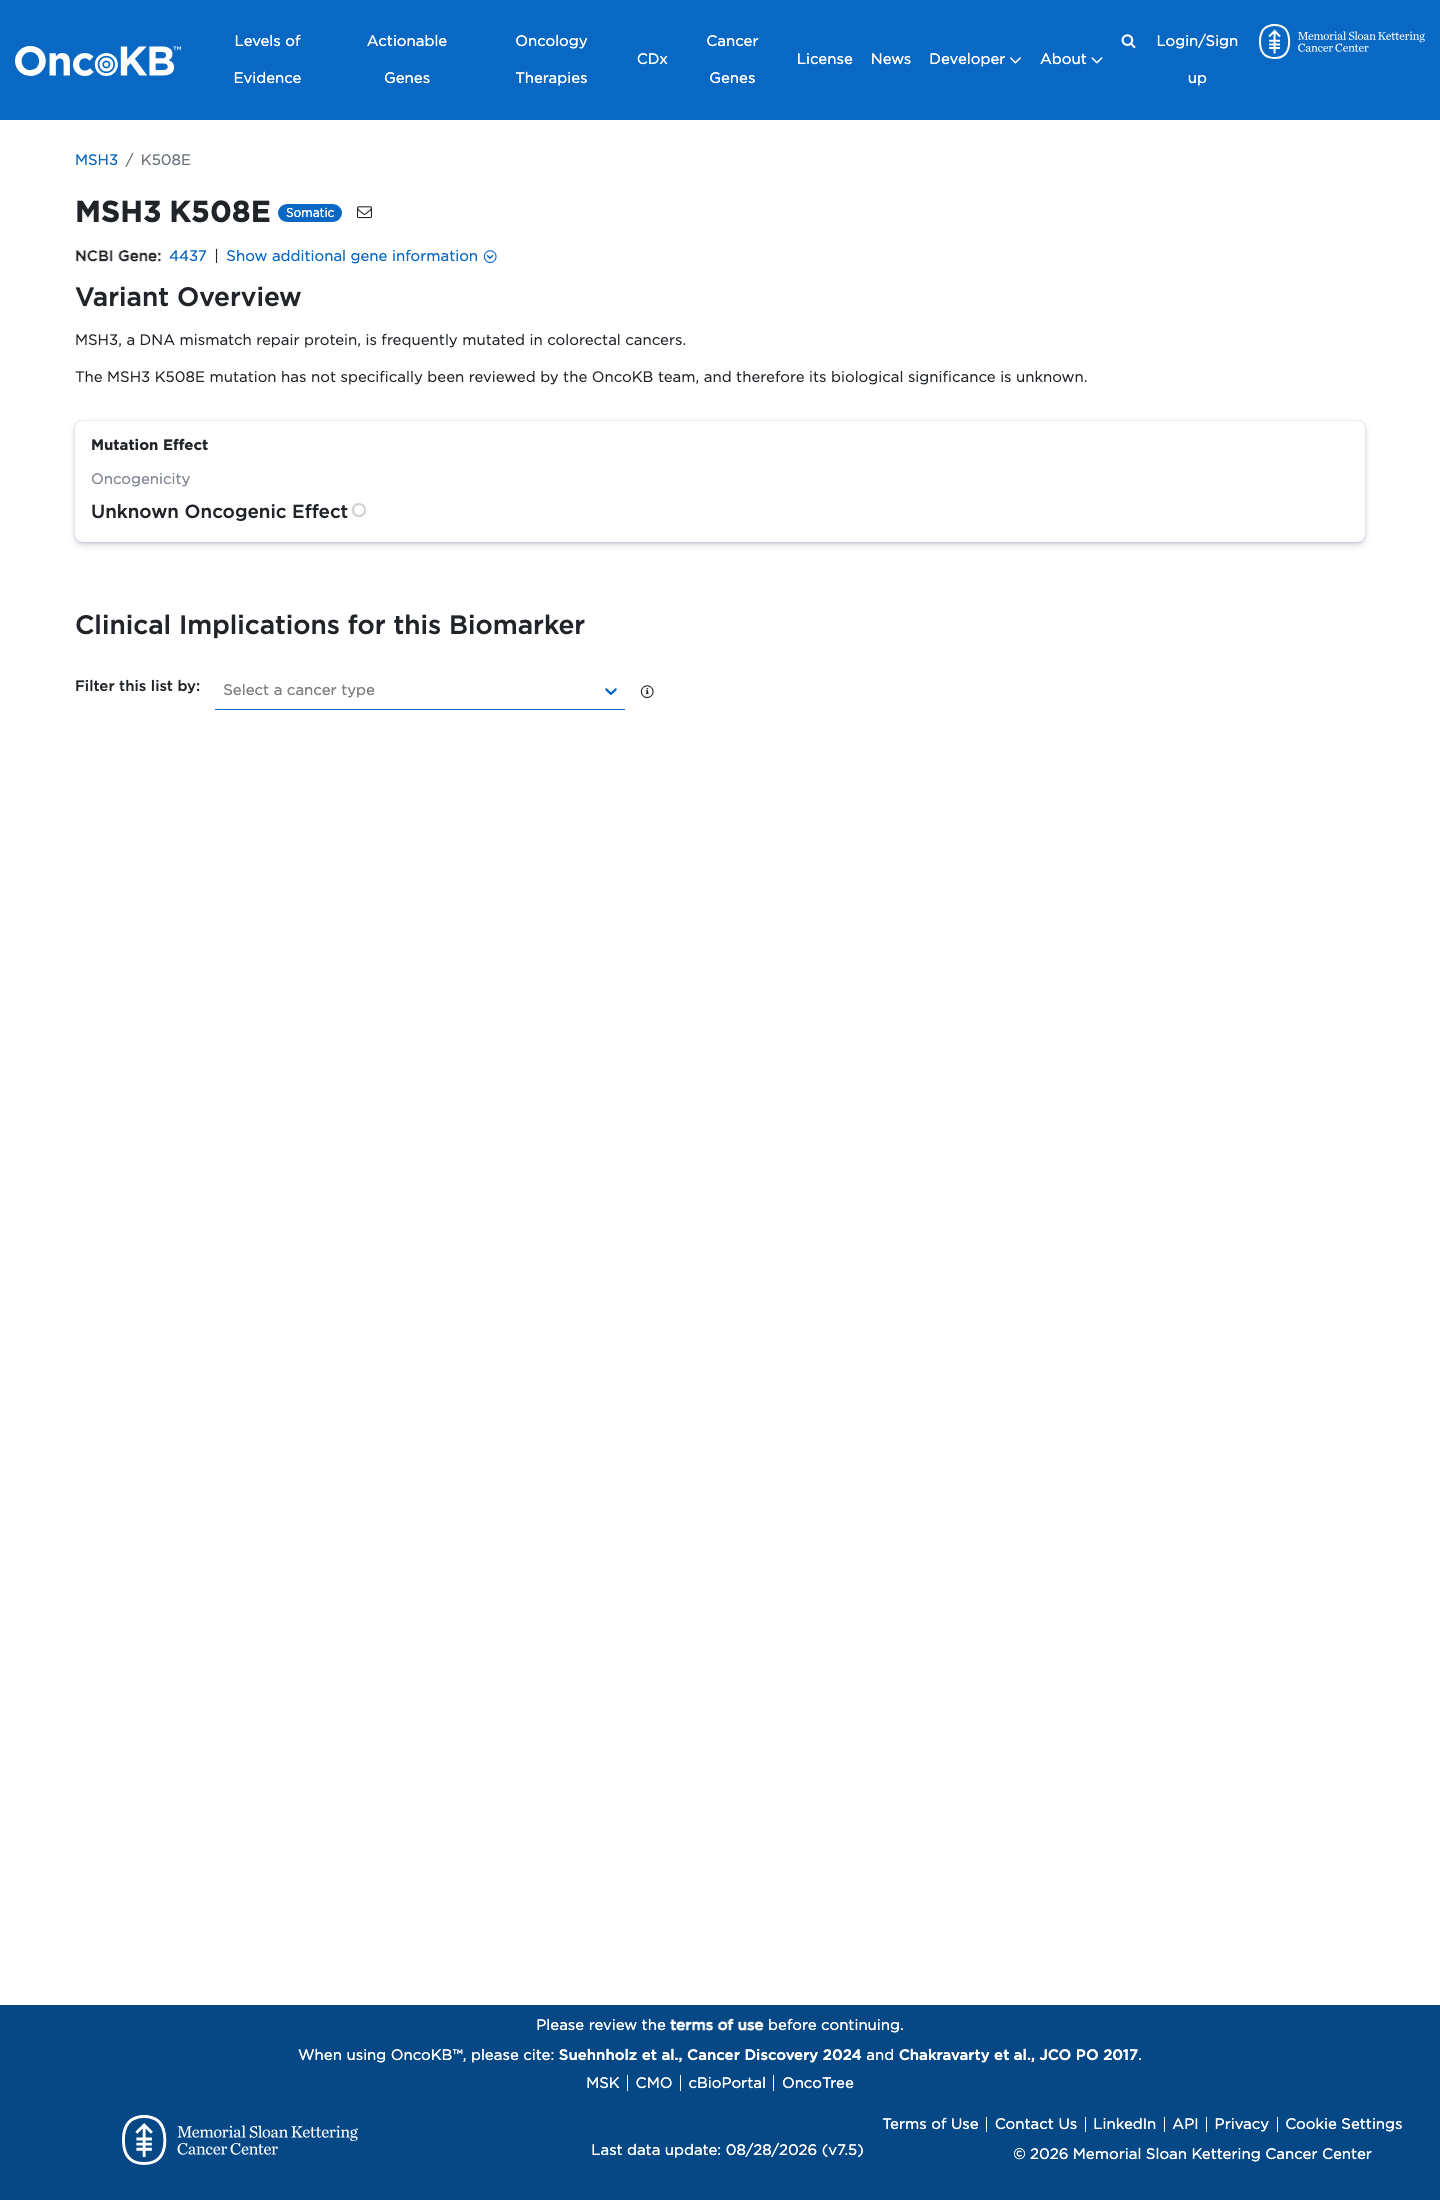
Functional / OncoKB screenshot
Functional Unknown Oncogenic Effect
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. MSH3, a DNA mismatch repair protein, is frequently mutated in colorectal cancers.
OncoKB ↗
COSMIC screenshot
COSMIC
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
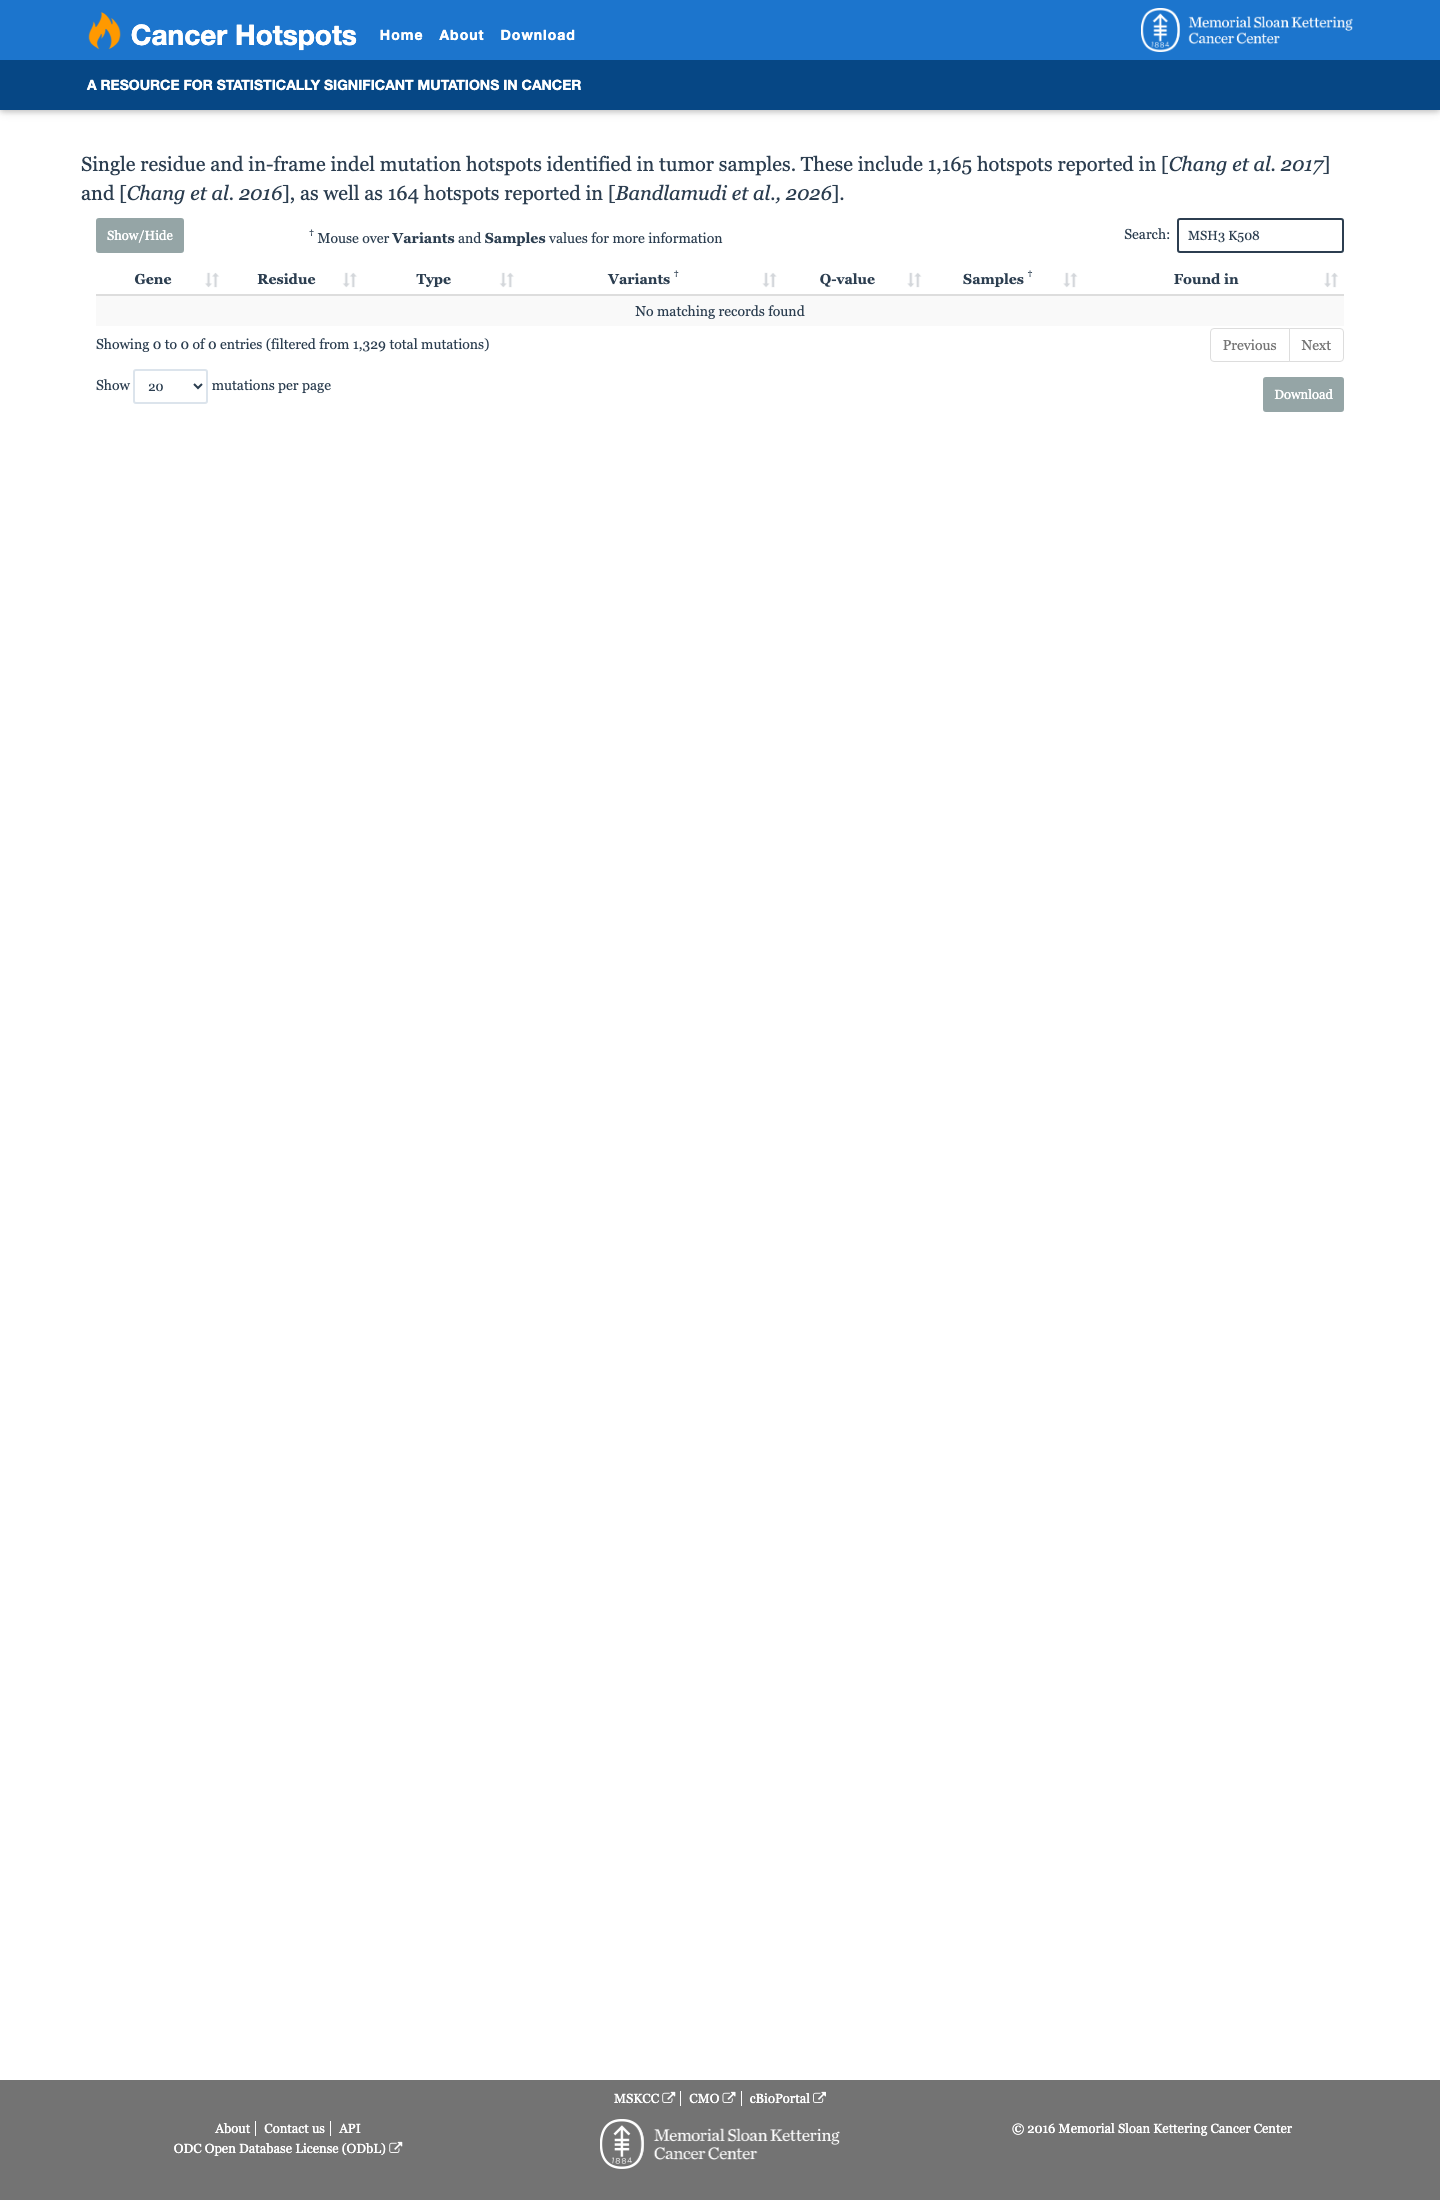
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
2papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 6 further PMIDs triaged but not cited — see Sources & references.
Rule & framework references · cited for criterion definitions, not variant evidence
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.
28944238 ↗ Targeted sequencing of 36 known or putative colorectal cancer susceptibility genes.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 6 PMIDs not cited in assessment
24493721 ↗ American Society of Clinical Oncology Expert Statement: collection and use of a cancer family history for oncology providers. CLINVAR
24929052 ↗ Endometrial cancer: a review and current management strategies: part II. CLINVAR
25394175 ↗ A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment. CLINVAR
26467025 ↗ A Standardized DNA Variant Scoring System for Pathogenicity Assessments in Mende CLINVAR
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification crite CLINVAR
33451724 ↗ Endometrial cancer: A society of gynecologic oncology evidence-based review and recommendations, part II. CLINVAR