PS1
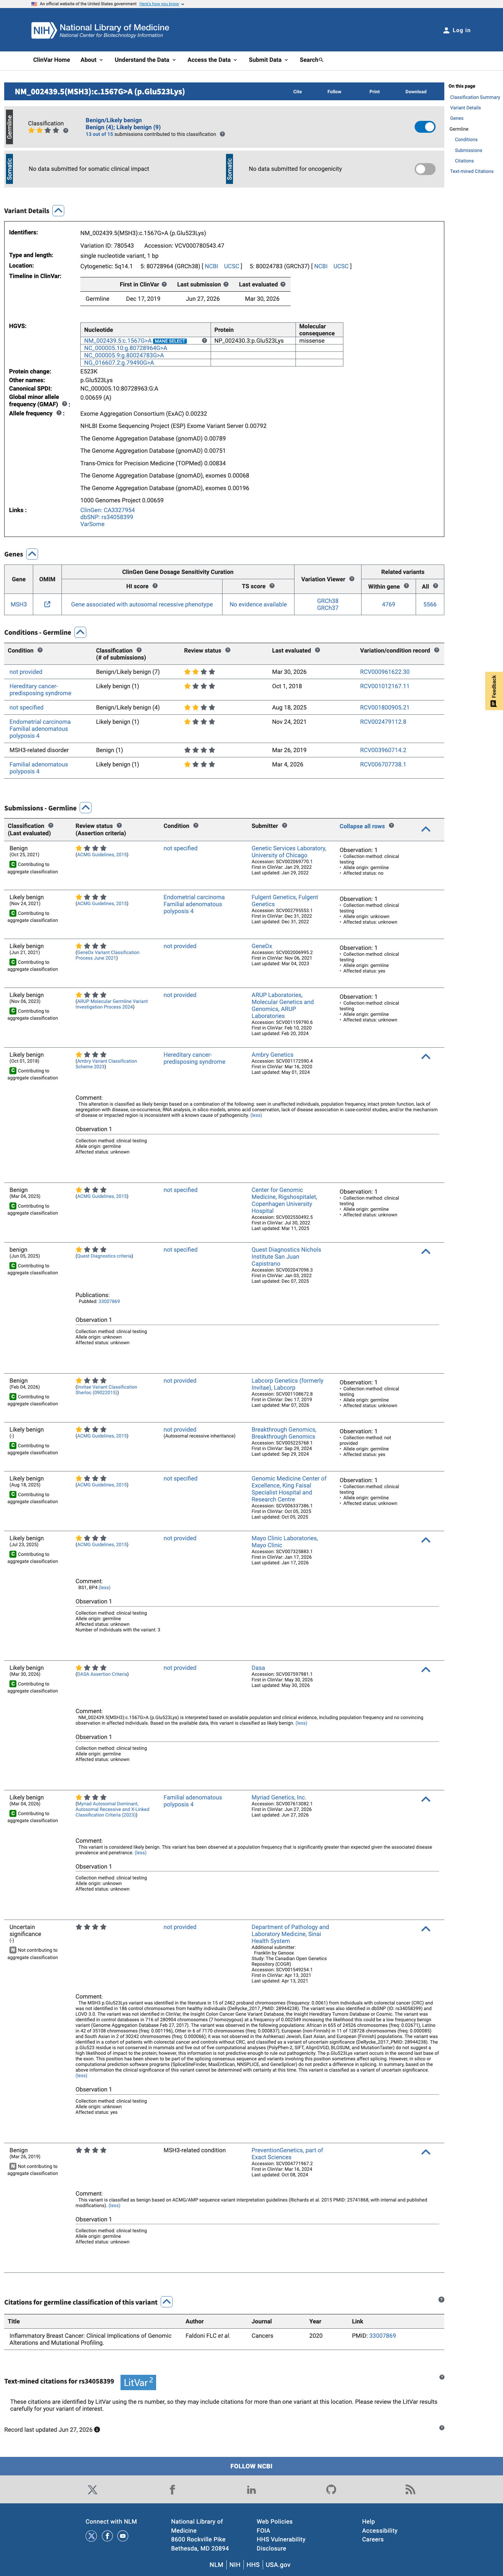
Not met: no established pathogenic variant carries p.Glu523Lys, and the only variant with this amino-acid change (ClinVar VCV000780543) is classified Benign/Likely benign by 15 submitters.
PS2
Not assessed: no proband, parental testing, or pedigree data were provided, so a confirmed de novo occurrence in an affected patient could not be evaluated.
PS3
Not met: no functional assay of MSH3 p.(Glu523Lys) exists; the sole paper reporting it gives no functional data.
PS4
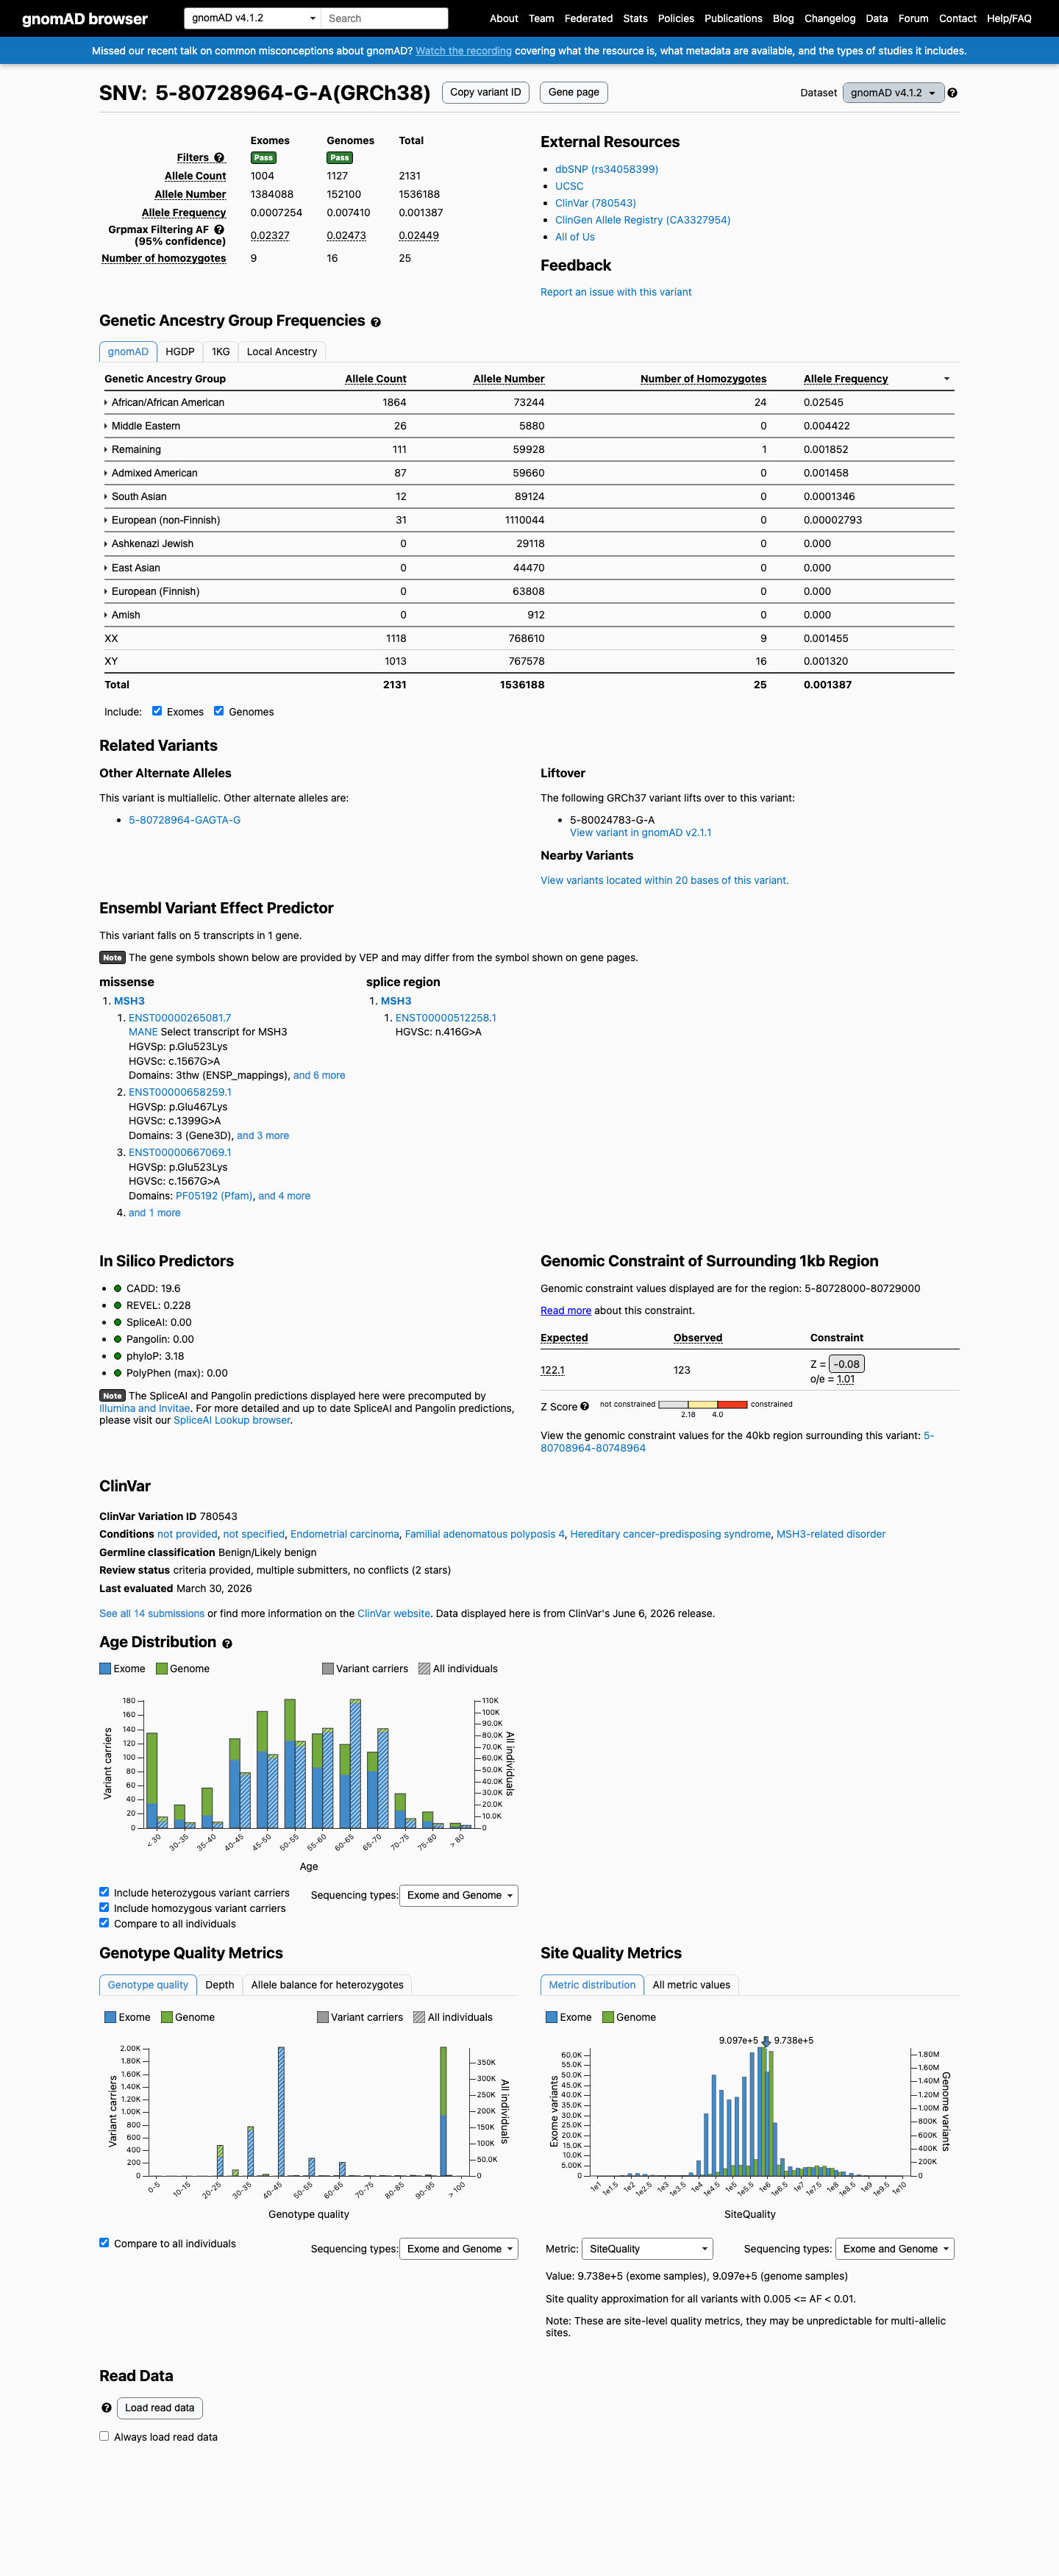
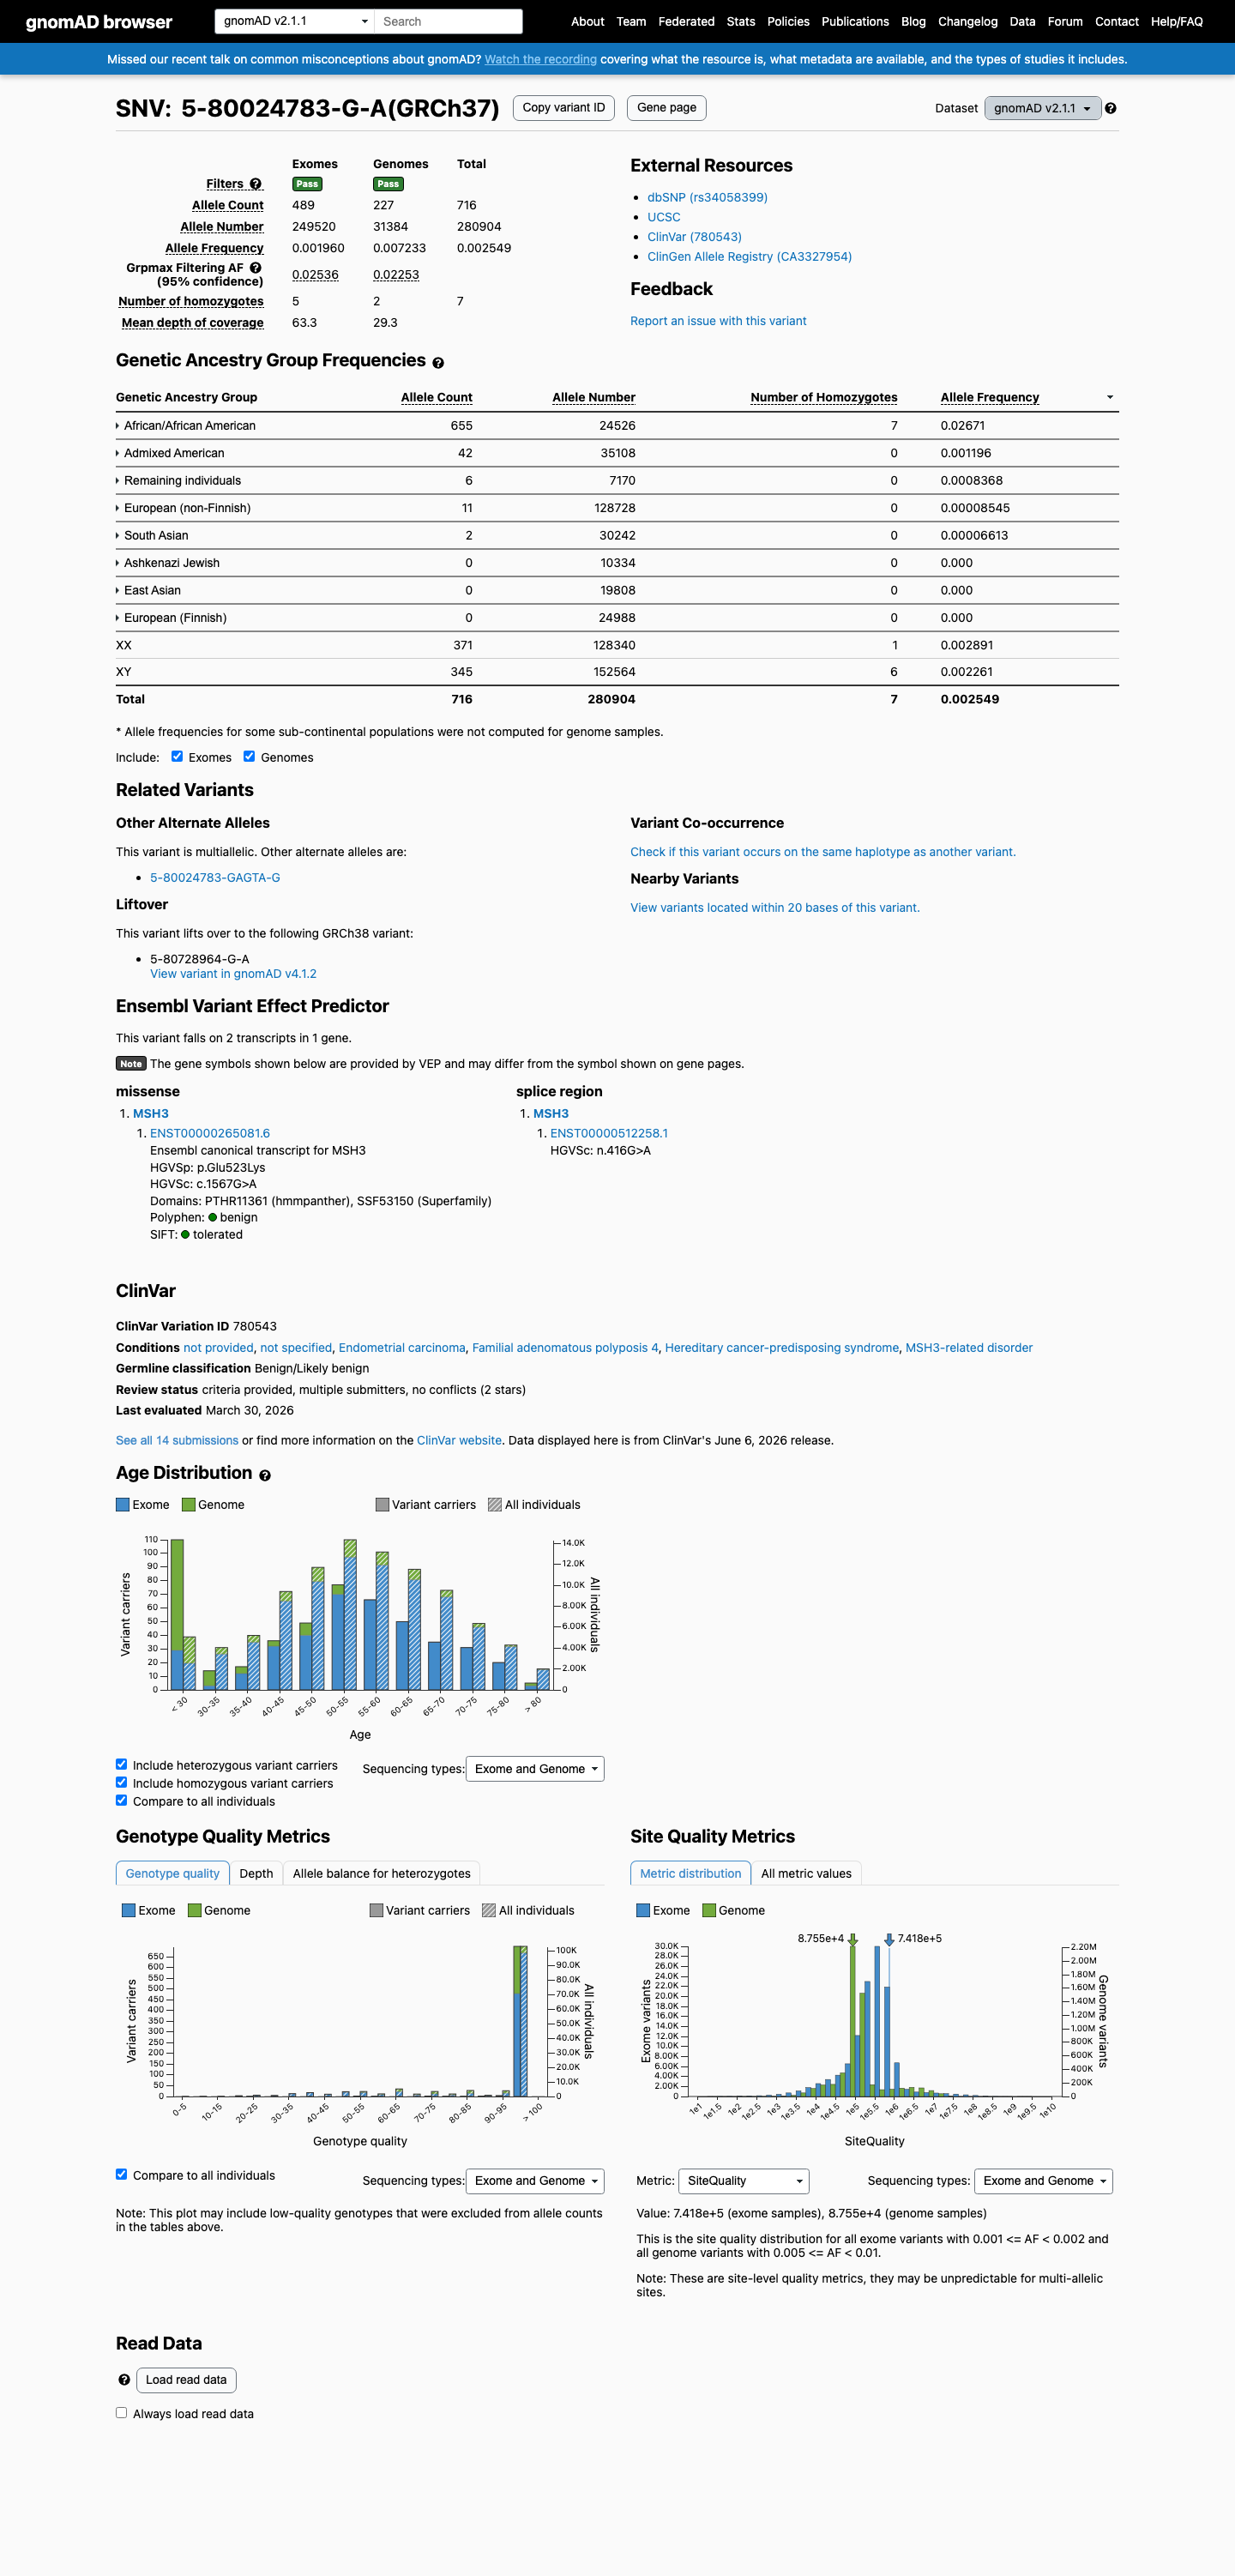
Not met: no case-control enrichment exists for c.1567G>A, which is instead common in gnomAD (popmax 2.5% African/African American).
PM1
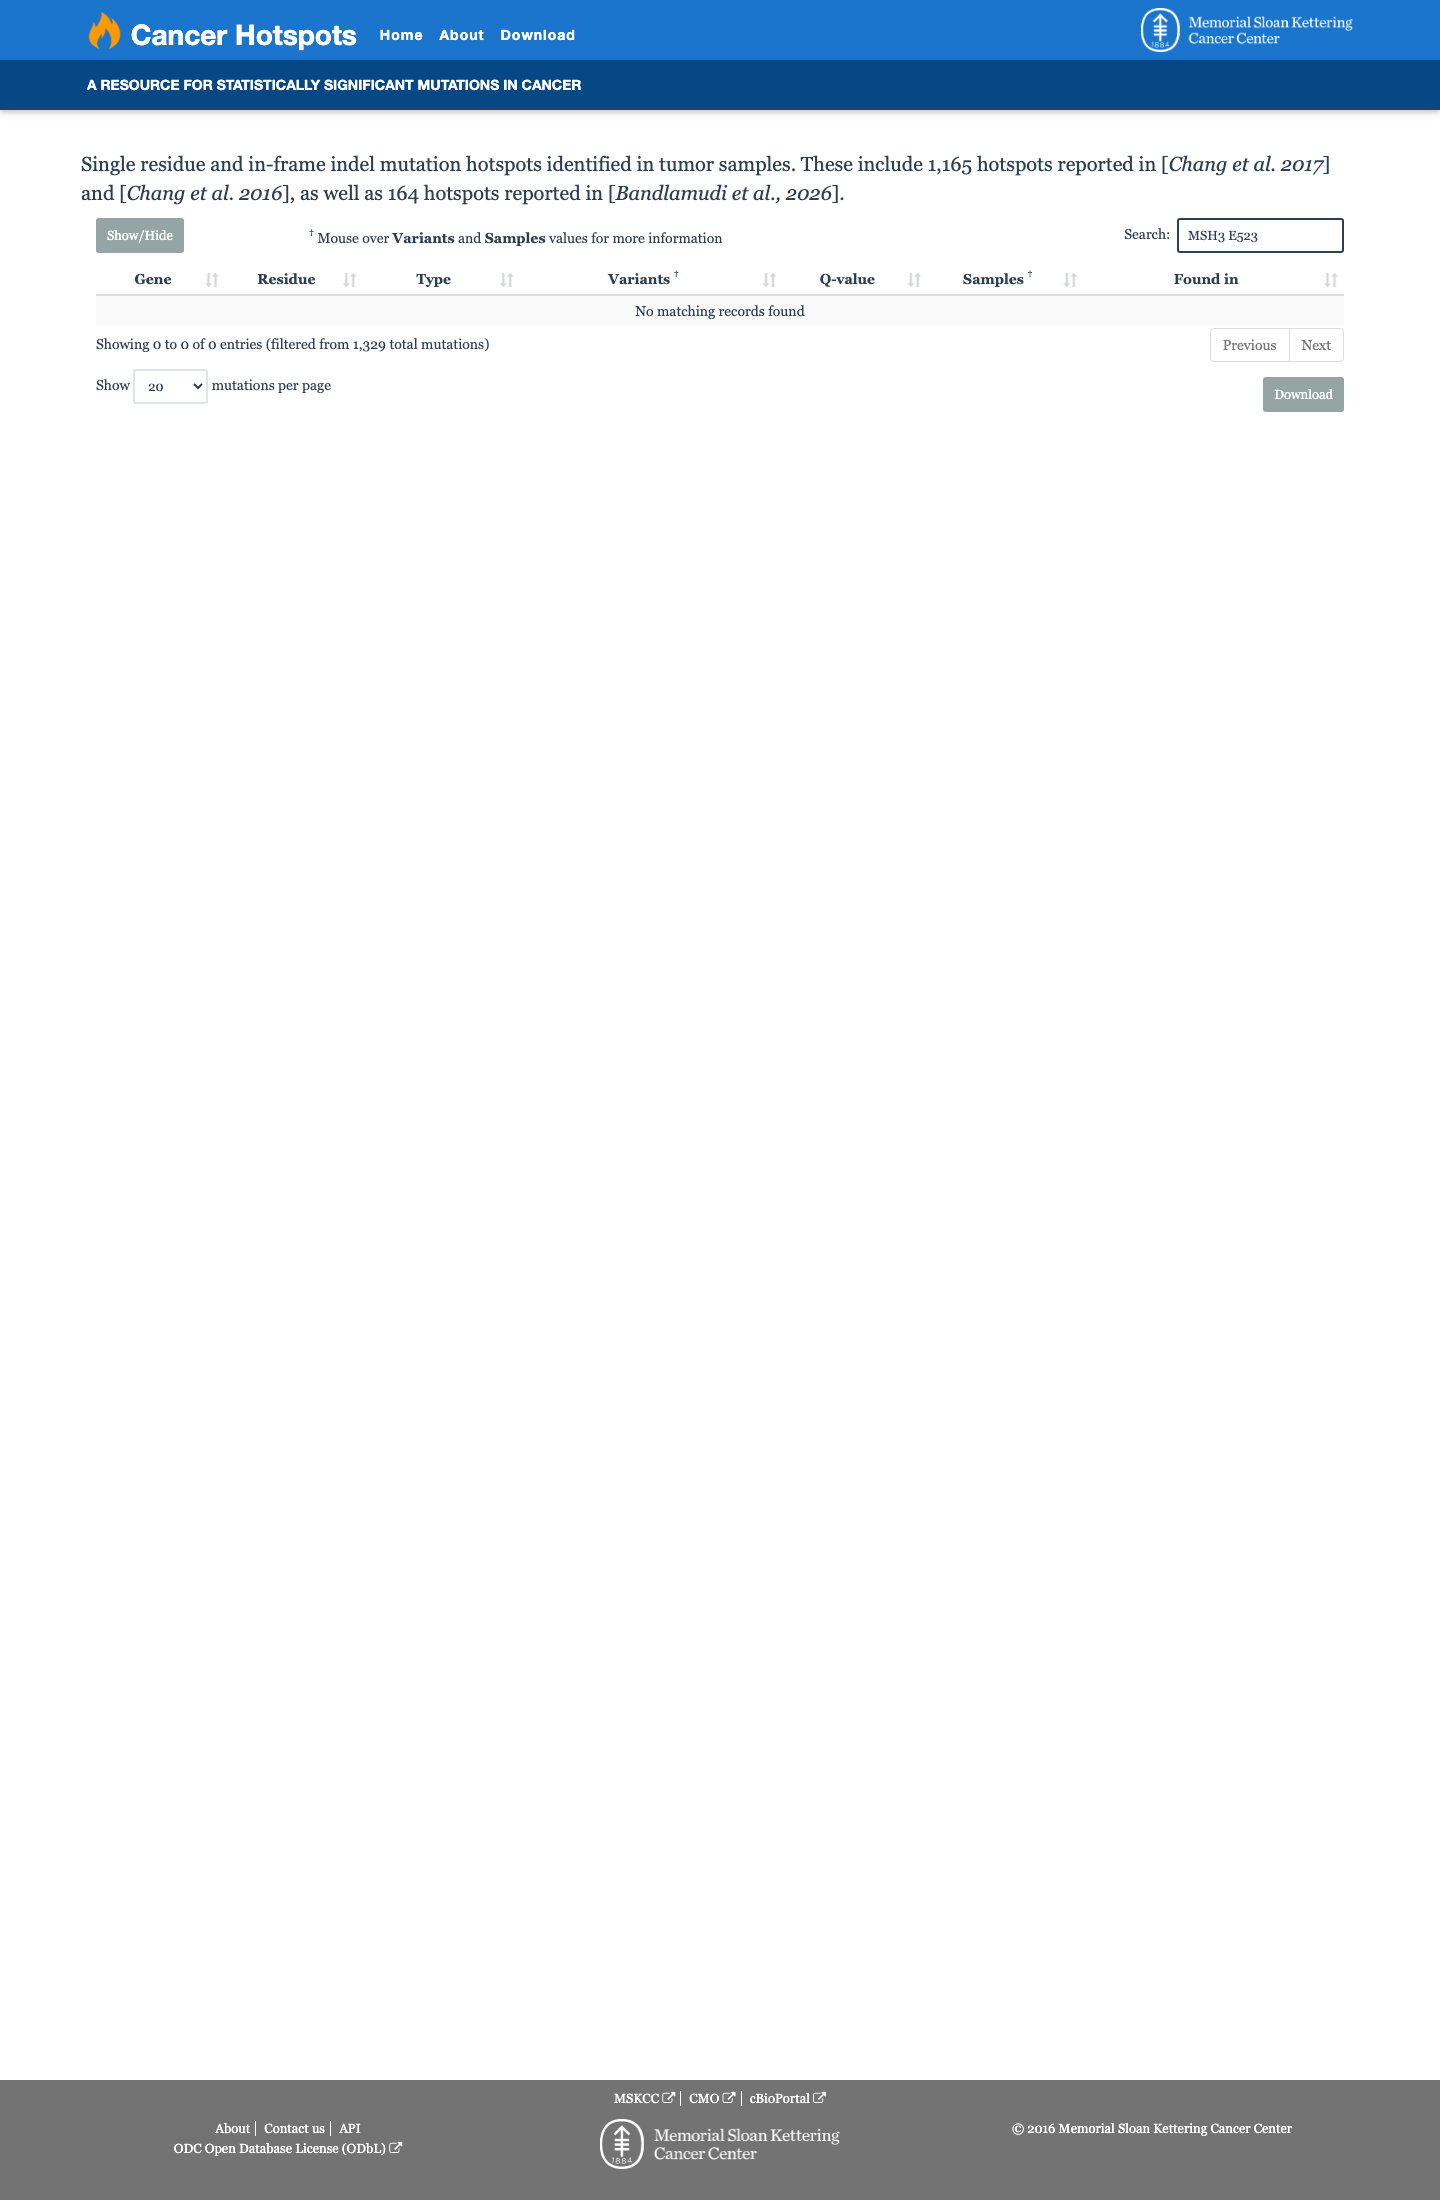
Not met: MSH3 E523 is not a significant hotspot (CancerHotspots: no result), lies outside any annotated functional domain, and carries gnomAD v4.1 AF 0.139% with 25 homozygotes.
PM2
Not met: gnomAD v4.1 allele frequency 0.0013872 (2,131/1,536,188 alleles) is about 14-fold above the 0.0001 PM2 supporting threshold.
PM3
Not met: no case pairs this allele in trans with a pathogenic MSH3 variant, and the only reported carrier (tumour NGS, case IBC15) has undetermined phase.
PM5
Not met: residue 523 has no established pathogenic missense, only p.Glu523Val as uncertain significance (single submitter) in ClinVar.
PM6
Not assessed: the record contains no parental or proband data suggesting an assumed de novo occurrence, so PM6 cannot be evaluated.
PP1
Not assessed: no pedigree, affected relatives, or co-segregation data were available for this variant in MSH3.
PP2
Not met: MSH3 tolerates missense variation (gnomAD oe_mis 1.083, mis_z -1.295) and ClinVar pathogenic MSH3 records are 480 truncating versus one missense.
PP3
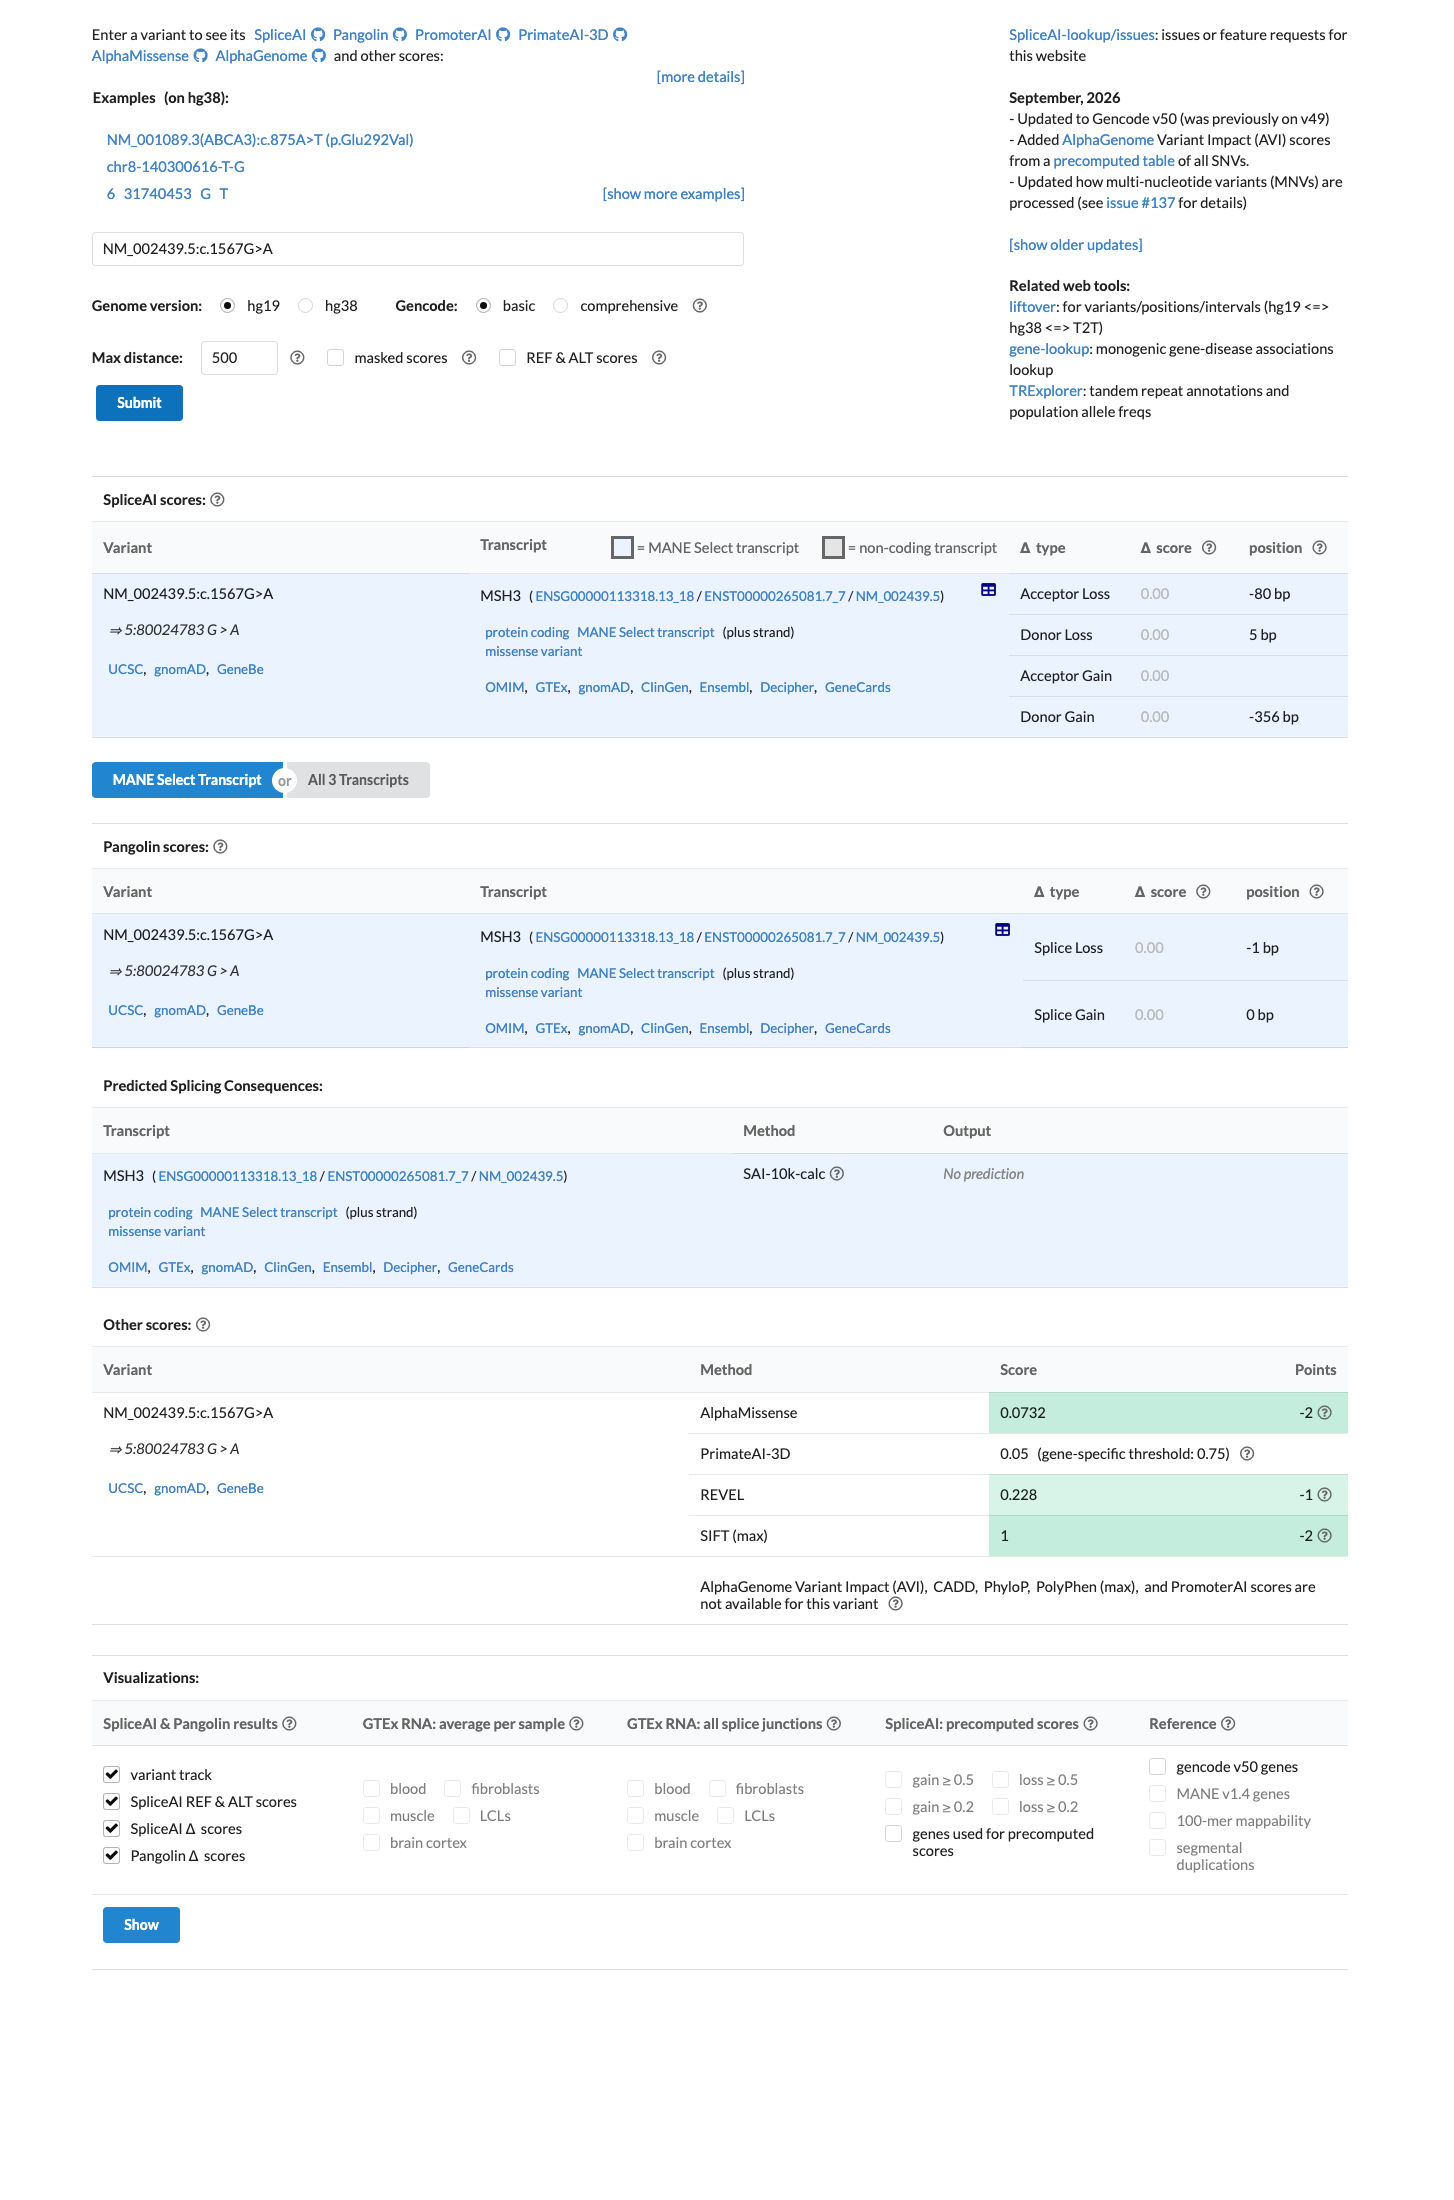
Not met: REVEL 0.228 is far below the >=0.644 supporting PP3 threshold for this missense variant.
PP4
Not assessed: no proband phenotype, HPO terms, or family history is available to test phenotype specificity.
PP5
Not met: ClinVar VCV000780543 has zero expert-panel submissions and only 2-star laboratory-level review for c.1567G>A.