Back
NM_000455.5:c.-1C>T
p.? · STK11
ACMG/AMP
0%
complete
Final classification
VUS
PM2BP4
STK11
c.-1C>T
p.?
canonical_splice · exon 1

STK11 encodes a serine/threonine kinase that acts as a tumor suppressor, regulating cell polarity, energy metabolism, and stress responses through the AMPK signaling pathway. Germline mutations in this gene cause Peutz-Jeghers syndrome, an inherited condition marked by gastrointestinal polyps, mucocutaneous pigmentation, and an elevated risk of several cancers. Loss of STK11 function also occurs in cancers of the lung, pancreas, breast, cervix, liver, and other tissues, where it promotes tumor growth.

This variant

STK11 encodes a serine/threonine kinase tumor suppressor whose germline loss of function causes autosomal dominant Peutz-Jeghers syndrome, so this 5' UTR substitution immediately upstream of the initiation codon was assessed for its potential to disrupt STK11 expression, and only rarity (PM2) and a computational prediction of no splice impact (BP4) could be established for it.
Transcript
NM_000455.5
HGVS · transcript:coding
NM_000455.5:c.-1C>T
GRCh38
chr19:1206913 C>T
GRCh37
chr19:1206912 C>T
VUS: PM2 (supporting) plus BP4 (supporting) form one supporting pathogenic against one supporting benign criterion, matching no ACMG/AMP 2015 combination rule.
Classification rationale
PM2 BP4 VUS
STK11 c.-1C>T canonical_splice · exon 1

PM2 (supporting): grpmax FAF 4.13e-06 (gnomAD v2.1), total AF 1.07e-05-2.10e-05 with zero homozygotes, well below the <=0.0001 rarity threshold. BP4 (supporting): SpliceAI max delta 0.01 on the splice path for this non-missense variant, below the <0.1 cut-off, predicting no impact on splicing. PVS1 (not applicable): c.-1C>T is a 5' UTR substitution upstream of the initiation codon and not a null class, so no very strong pathogenic evidence applies. PS4, PP4, BP5 (not met): no case-control enrichment, no proband phenotype, and no alternative molecular cause are documented for this variant. BA1/BS1/BS2 (not met): maximum population frequency 1.15e-04 is far below the benign frequency thresholds and no homozygotes were observed. PP5/BP6 (not met): ClinVar record 185127 is laboratory-only with 0 expert-panel submissions and conflicting 1-star classifications.

PM2 + BP4 → VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_000455.5 · variants mapped to exon structure
STK11 NM_000455.5
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 16 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
PM2 supporting Pathogenic
Met (supporting): grpmax FAF 4.13e-06 (gnomAD v2.1) is well below the <=0.0001 PM2 rarity threshold.
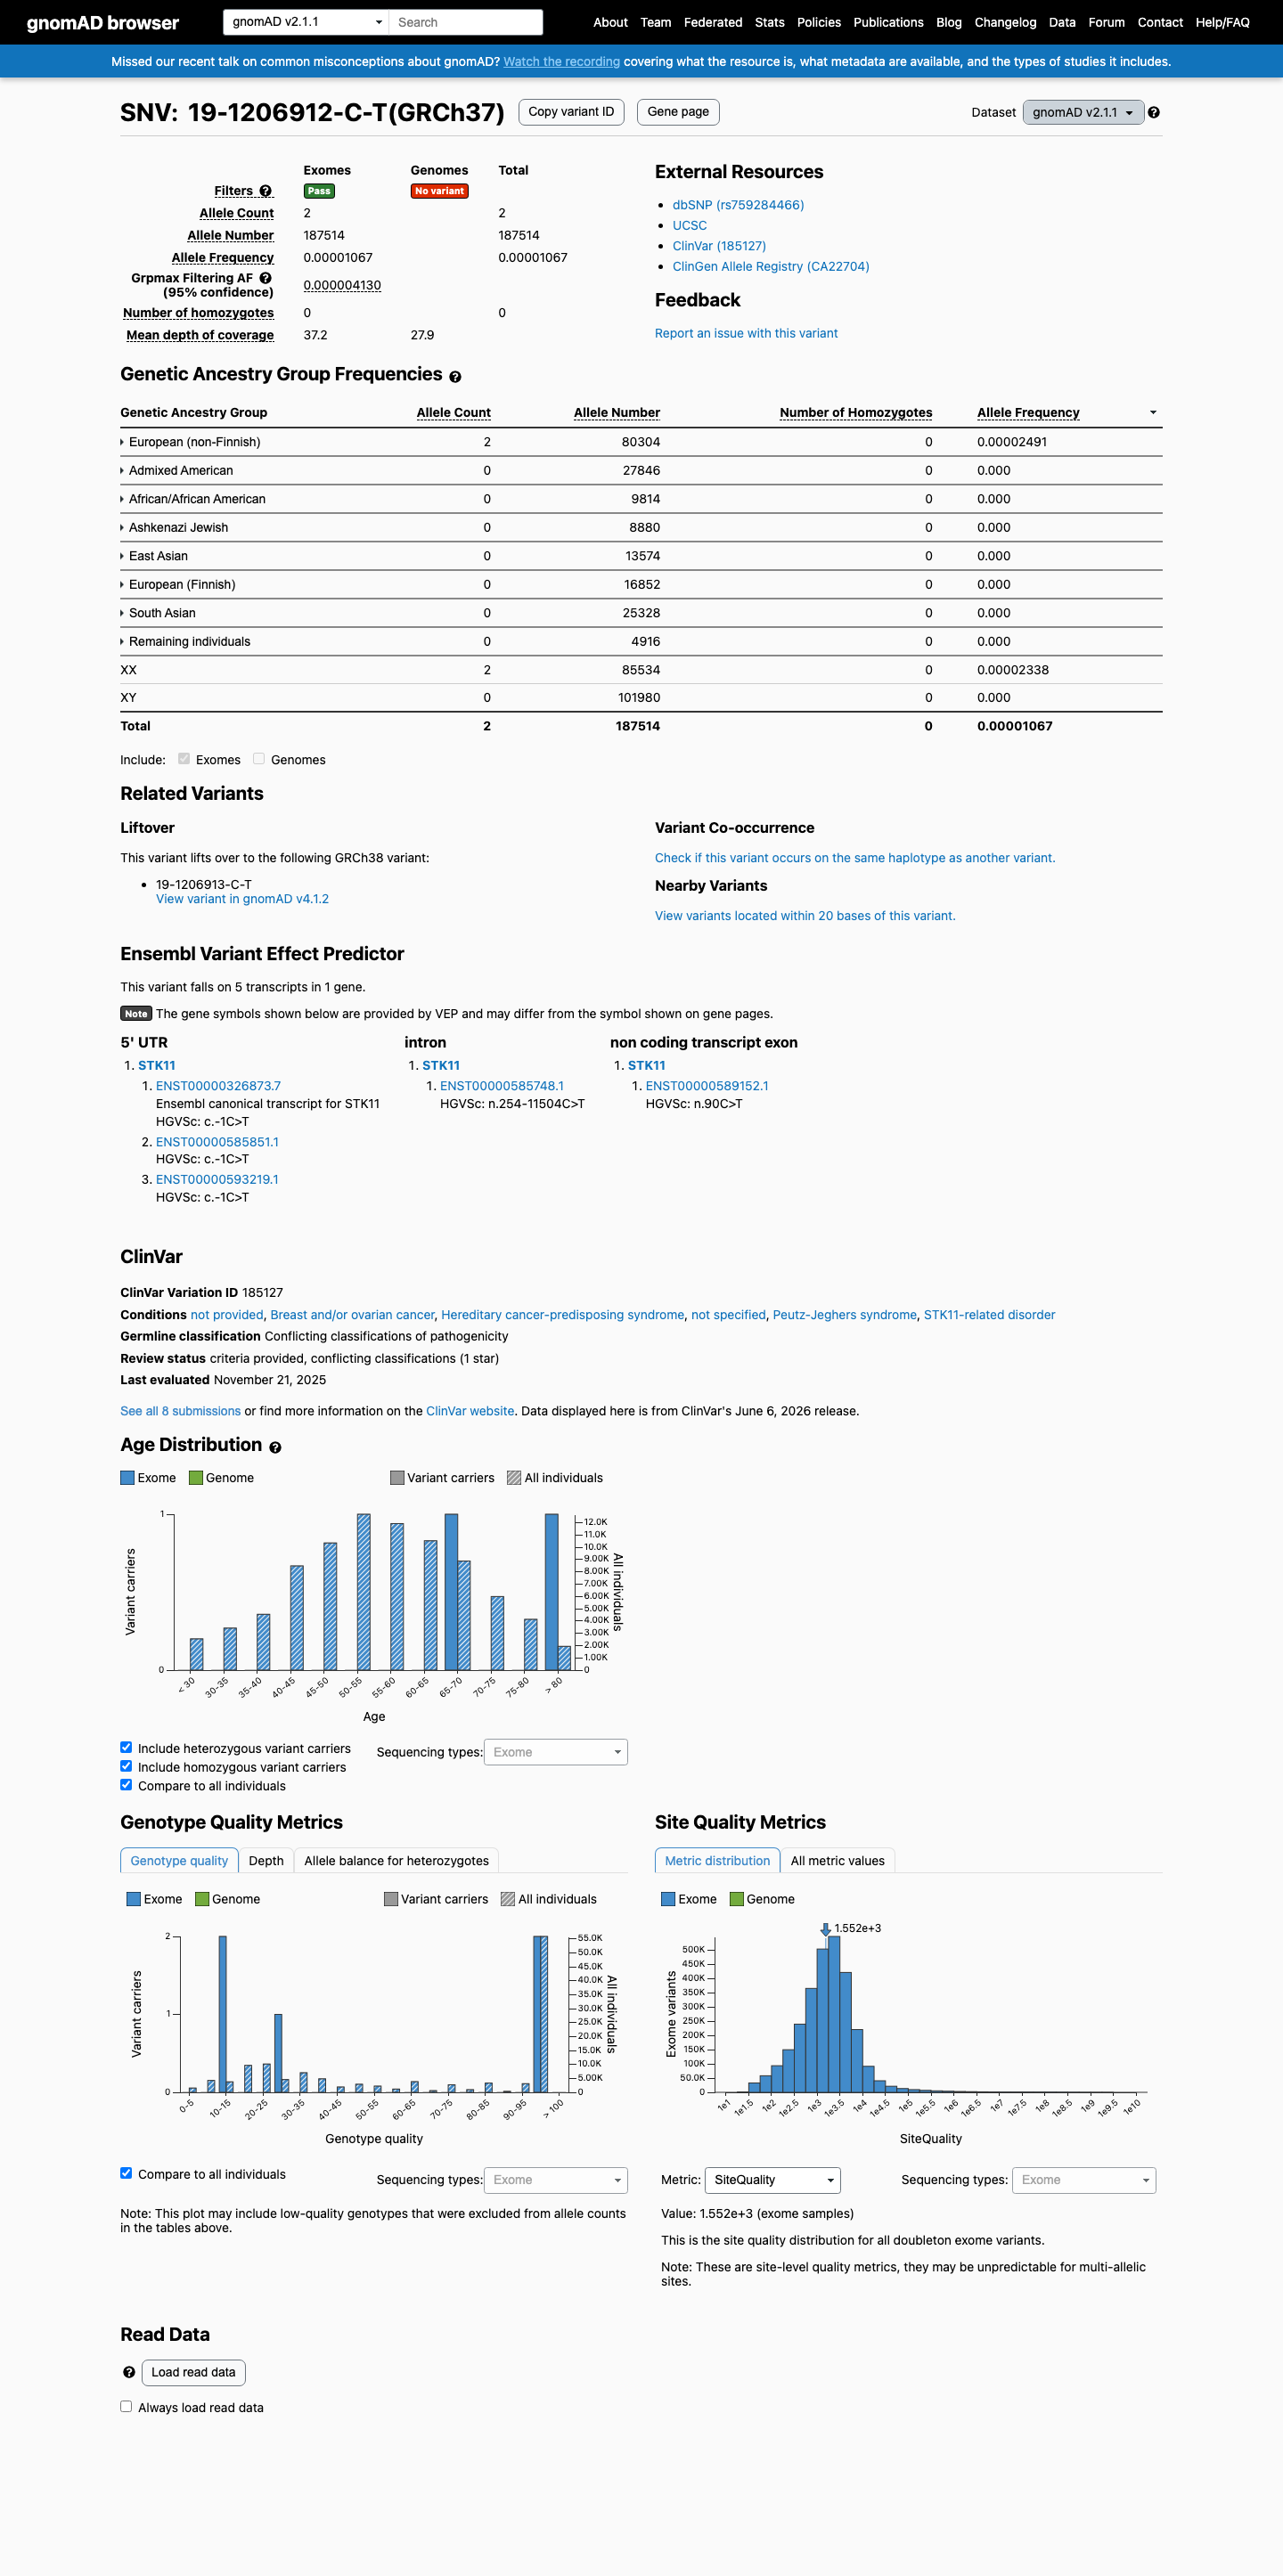
gnomAD v2.1 (all-comers): total AF 1.06659e-05 (2/187514), grpmax FAF 4.13e-06, homozygotes 0.gnomAD v4.1 (all-comers): total AF 2.09512e-05 (33/1575088), grpmax FAF 1.558e-05, homozygotes 0; highest population 'Remaining individuals' 1.14664e-04.gnomAD v2.1 non-cancer exomes: AF 1.10894e-05 (2/180352), homozygotes 0.
BP4 supporting Benign
Met (supporting): SpliceAI max delta 0.01 versus the <=0.1 BP4 supporting threshold.
SpliceAI lookup for NM_000455.5:c.-1C>T (hg37) returned max delta score 0.01, at or below the <=0.1 BP4 supporting cutoff (Jaganathan et al. 2019, Cell; PMID:30661751).BP4 taken from the SpliceAI (splice) path only, matched to the variant's canonical_splice consequence class; no evidence was drawn from the REVEL/BayesDel missense path.No ClinGen CSPEC or VCEP/gene-specific PP3/BP4 lookup specification exists for STK11 (vcep_materials.json found=false; framework_mode generic_acmg), so no pre-assigned VCEP code overrides the generic cutoff.
Assessed · not applied · 9 not met · 7 not assessed
Pathogenic
PS2 Not assessed: zero probands with confirmed parental testing are documented, so a de novo occurrence of c.-1C>T cannot be established.
PS3 Not assessed: no minigene, transcript, or protein functional assay has been reported for STK11 c.-1C>T to evaluate a damaging effect.
PS4 Not met: no case-control enrichment data exist for this variant, and the only STK11 case series does not name c.-1C>T.
PM6 Not assessed: no affected individual with a presumed de novo c.-1C>T and unconfirmed parental testing is documented.
PP1 Not assessed: zero informative meioses and no variant-carrying affected relatives are documented for c.-1C>T.
PP3 Not met: SpliceAI max delta 0.01, far below the >=0.2 supporting PP3 threshold.
PP4 Not met: no proband phenotype or family history is available to demonstrate disease specificity for this variant.
PP5 Not met: ClinVar record 185127 has 0 expert-panel submissions and a conflicting, 1-star laboratory-only review status.
Benign
BA1 Not met: maximum population frequency 1.15e-04 (gnomAD v4.1, Remaining individuals) is ~435-fold below the 0.05 BA1 stand-alone threshold.
BS1 Not met: highest subpopulation frequency 1.15e-04 (gnomAD v4.1, Remaining individuals) is ~87-fold below the 0.01 BS1 threshold.
BS2 Not met: zero homozygotes across all gnomAD datasets, and the adult-onset, incompletely penetrant autosomal dominant Peutz-Jeghers phenotype does not satisfy BS2's requirement of full penetrance at an early age.
BS3 Not assessed: no functional assay reporting normal STK11 c.-1C>T splicing or protein function is available to evaluate BS3.
BS4 Not assessed: no family genotypes are available, so non-segregation of c.-1C>T cannot be demonstrated.
BP2 Not assessed: no second STK11 variant or phasing data exist to establish whether this variant lies in cis or in trans.
BP5 Not met: no case carrying this variant has a documented alternative molecular cause.
BP6 Not met: ClinVar record 185127 contains 0 expert-panel submissions; its benign labels are ordinary laboratory assertions.
N/A · 10 PVS1 · PS1 · PM1 · PM3 · PM4 · PM5 · PP2 · BP1 · BP3 · BP7
Research & evidence
Population frequency
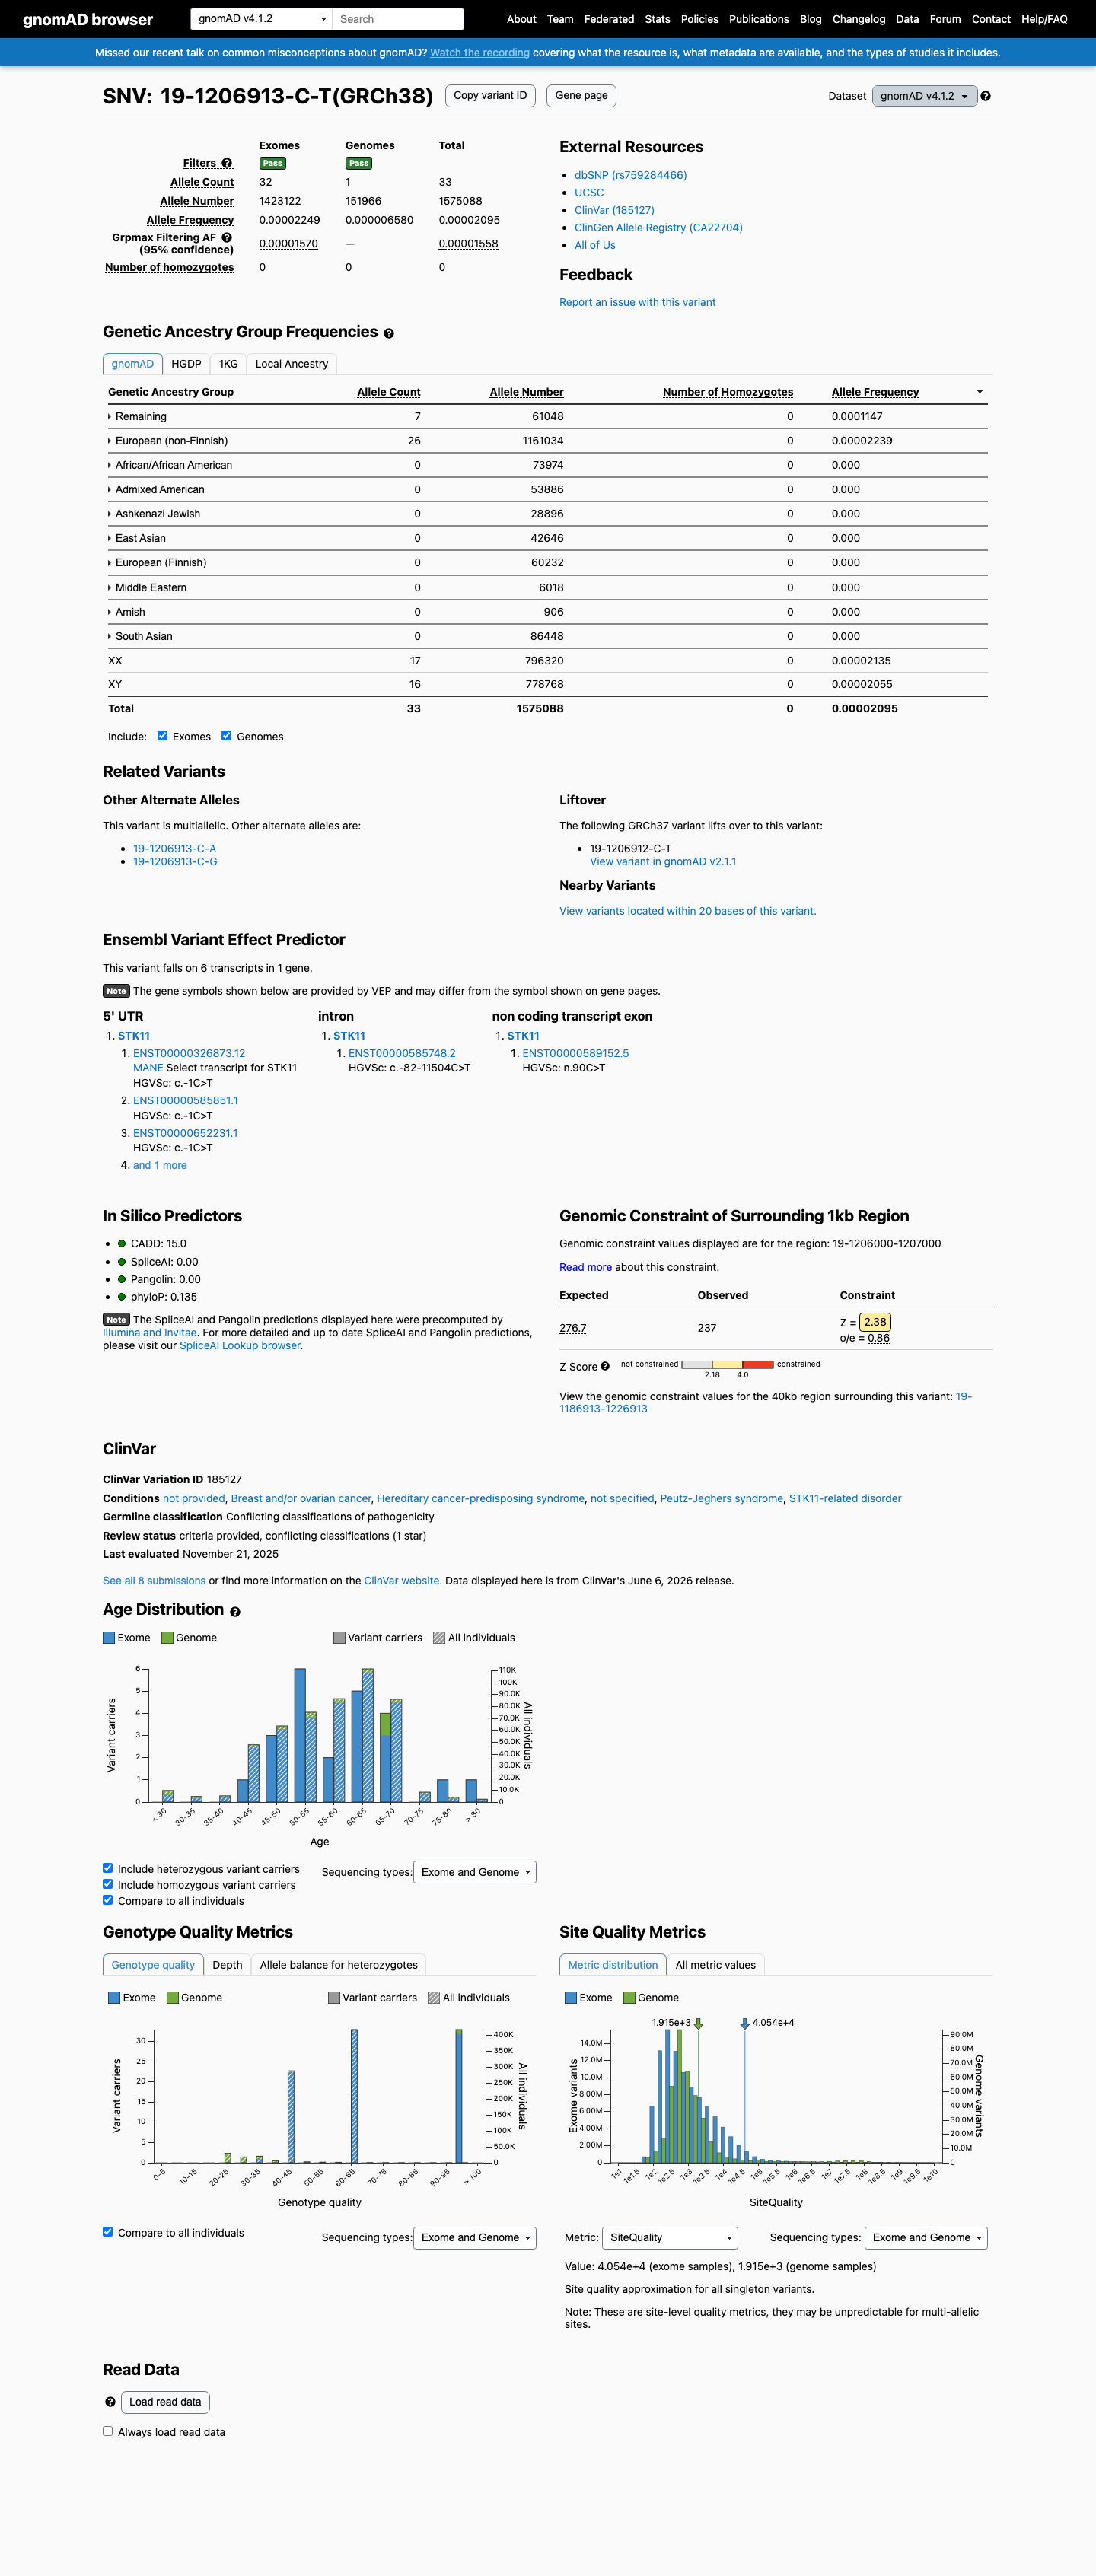
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 2.09512e-05; MAF= 0.00210%, 33/1575088 alleles, homozygotes = 0) and has highest observed frequency in the Remaining individuals population (AF= 0.000114664; MAF= 0.01147%, 7/61048 alleles, homozygotes = 0); grpmax FAF= 1.558e-05.
v2.1
This variant is present in gnomAD v2.1 (AF= 1.06659e-05; MAF= 0.00107%, 2/187514 alleles, homozygotes = 0) and has highest observed frequency in the European (non-Finnish) population (AF= 2.49054e-05; MAF= 0.00249%, 2/80304 alleles, homozygotes = 0); grpmax FAF= 4.13e-06.
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.0021% · 33 / 1,575,088
0 hom · FAF 0.0016%
Remaining individuals
7 / 61,048
0.011%
European (non-Finnish)
26 / 1,161,034
0.0022%
+ 8 not observed (Admixed American, European (Finnish), Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American)
gnomAD v2.1
0.0011% · 2 / 187,514
0 hom · FAF 0.00041%
European (non-Finnish)
2 / 80,304
0.0025%
+ 7 not observed (African/African American, Admixed American, Ashkenazi Jewish, East Asian, European (Finnish), Remaining individuals, South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
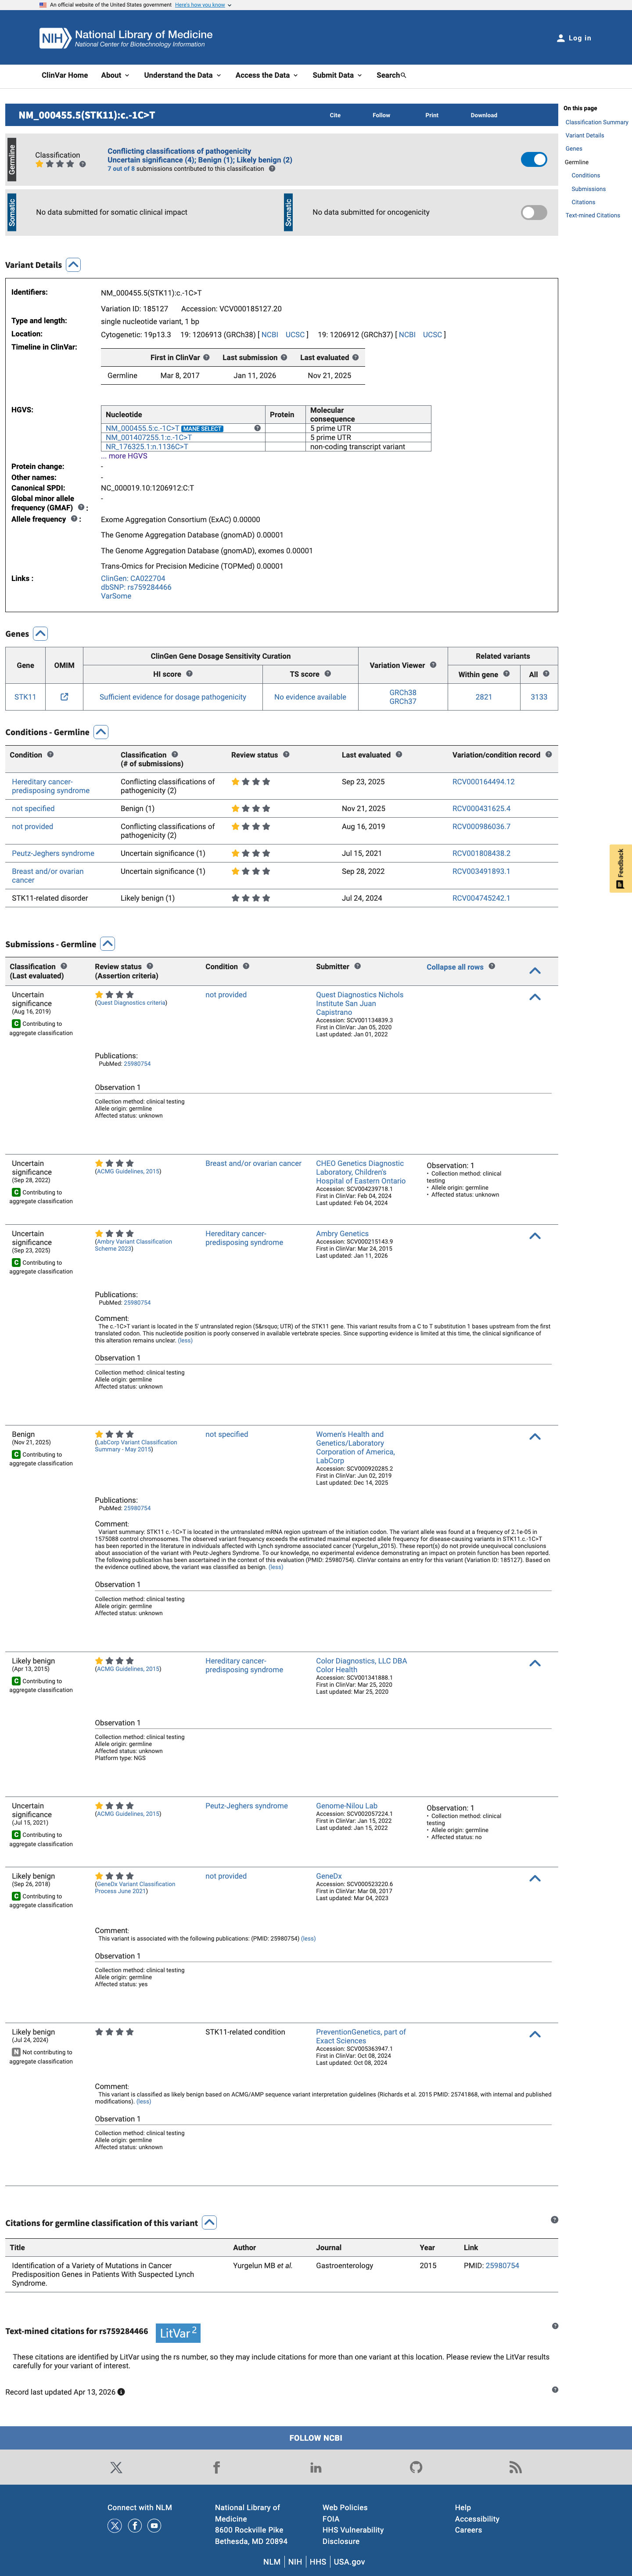
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Uncertain significance (4 clinical laboratories) and as Likely benign (3 clinical laboratories) and as Benign (1 clinical laboratory). (ClinVarID = 185127)
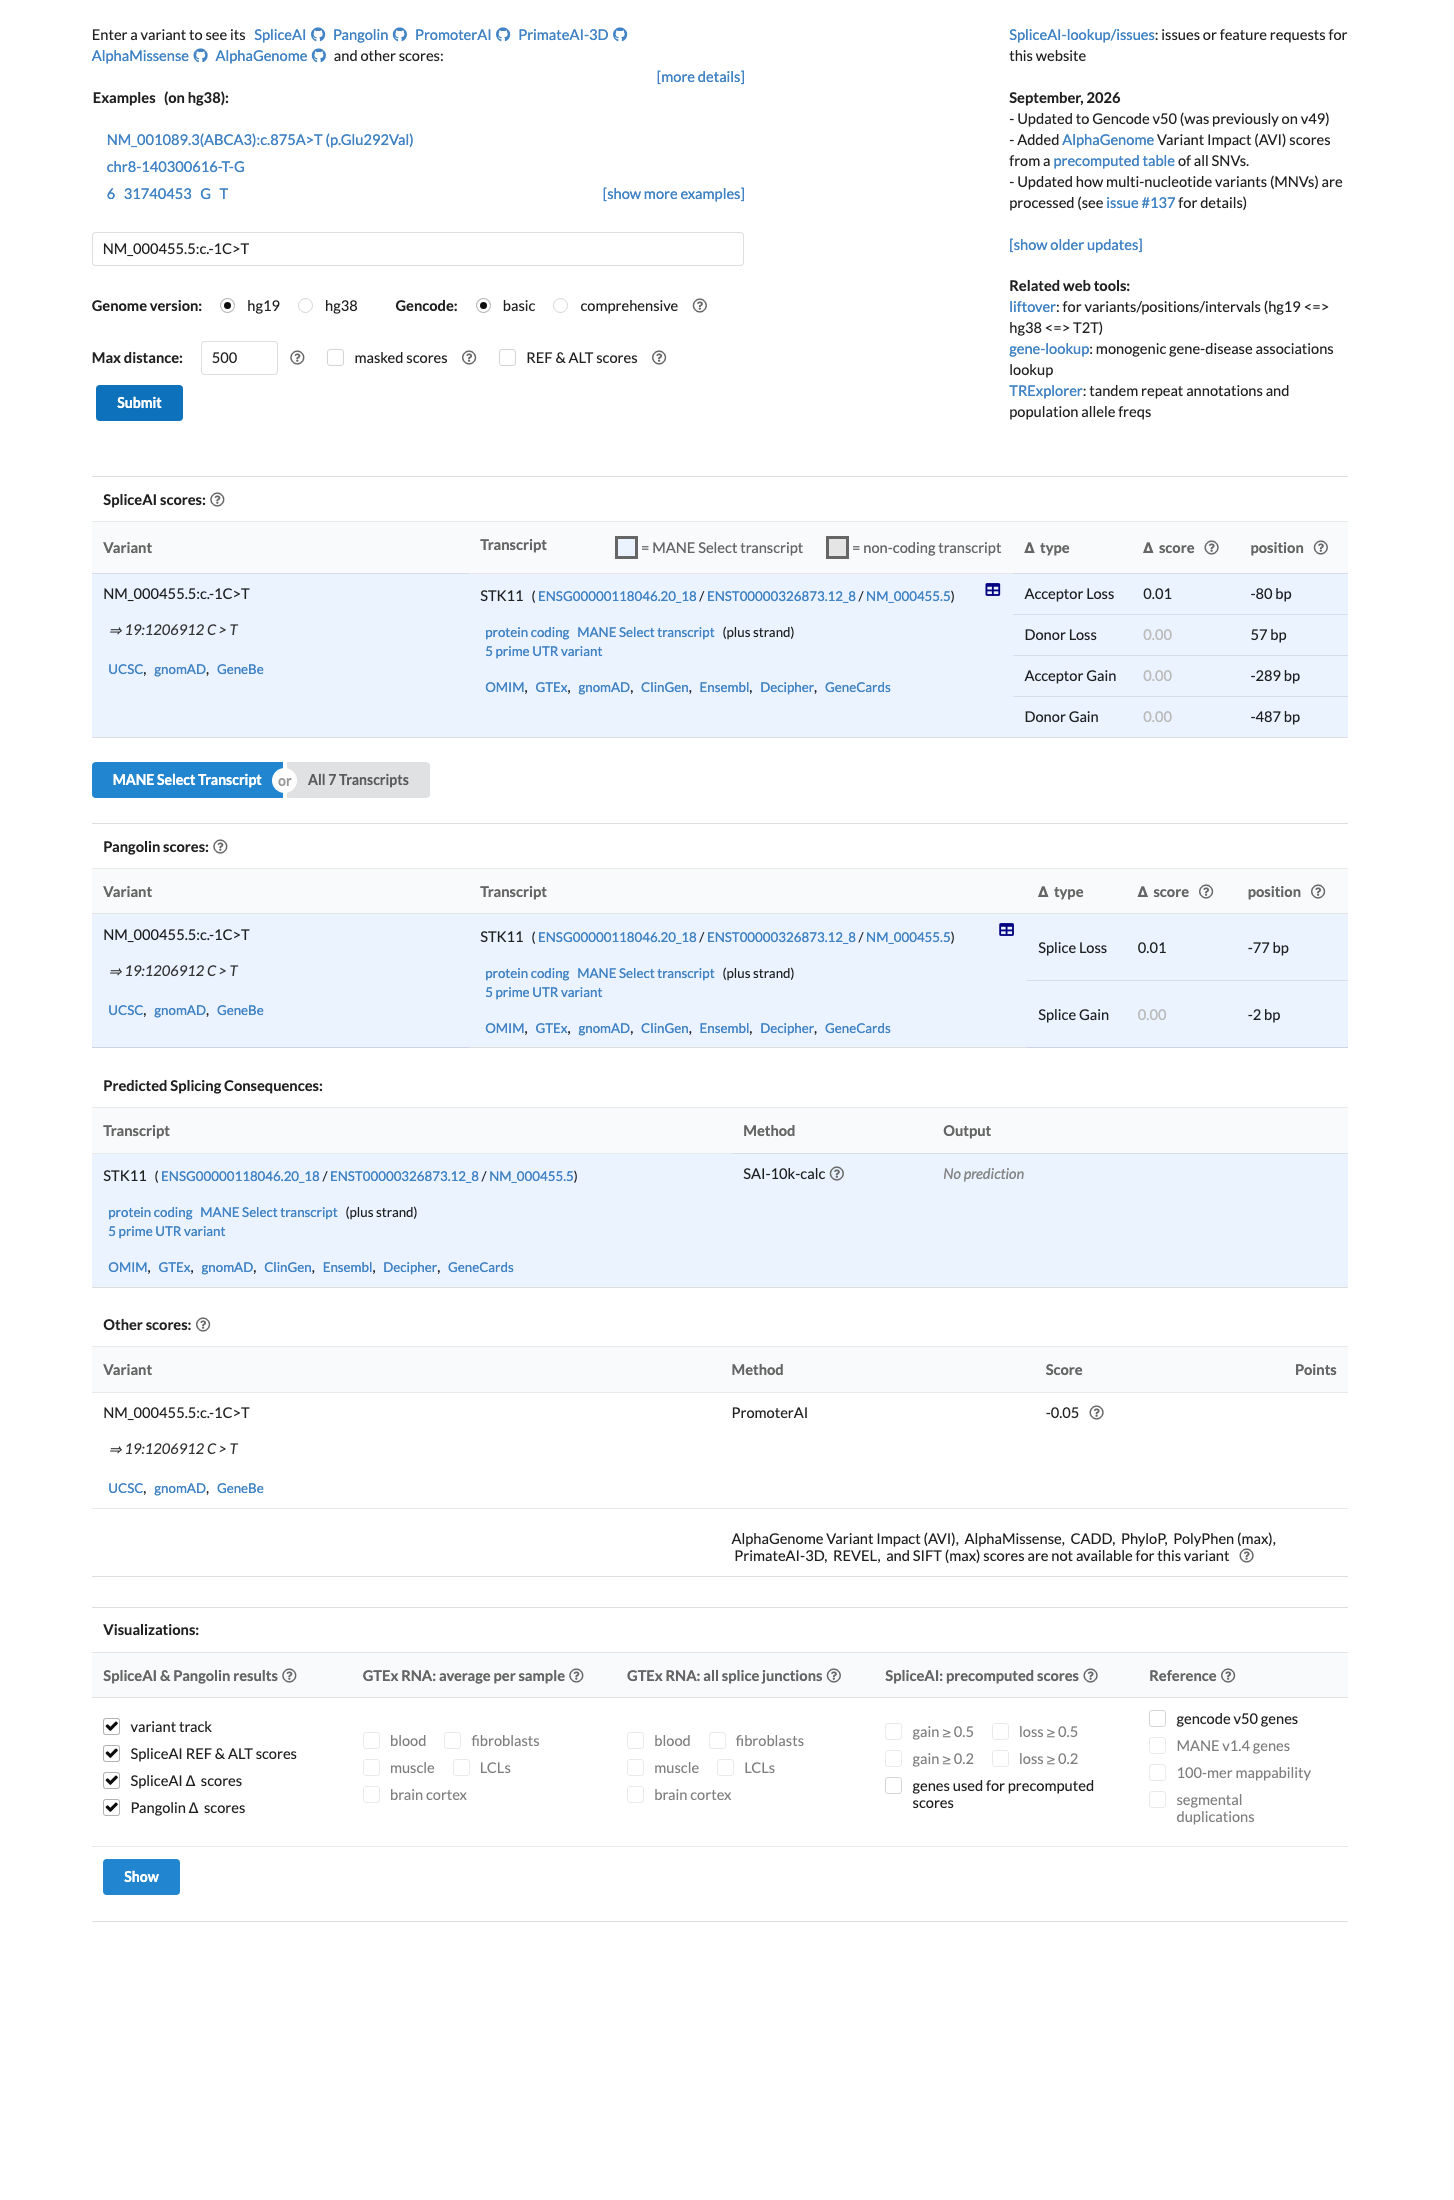
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.01).
Functional No data
No calibrated functional assay or RNA evidence was identified for this variant.
OncoKB ↗
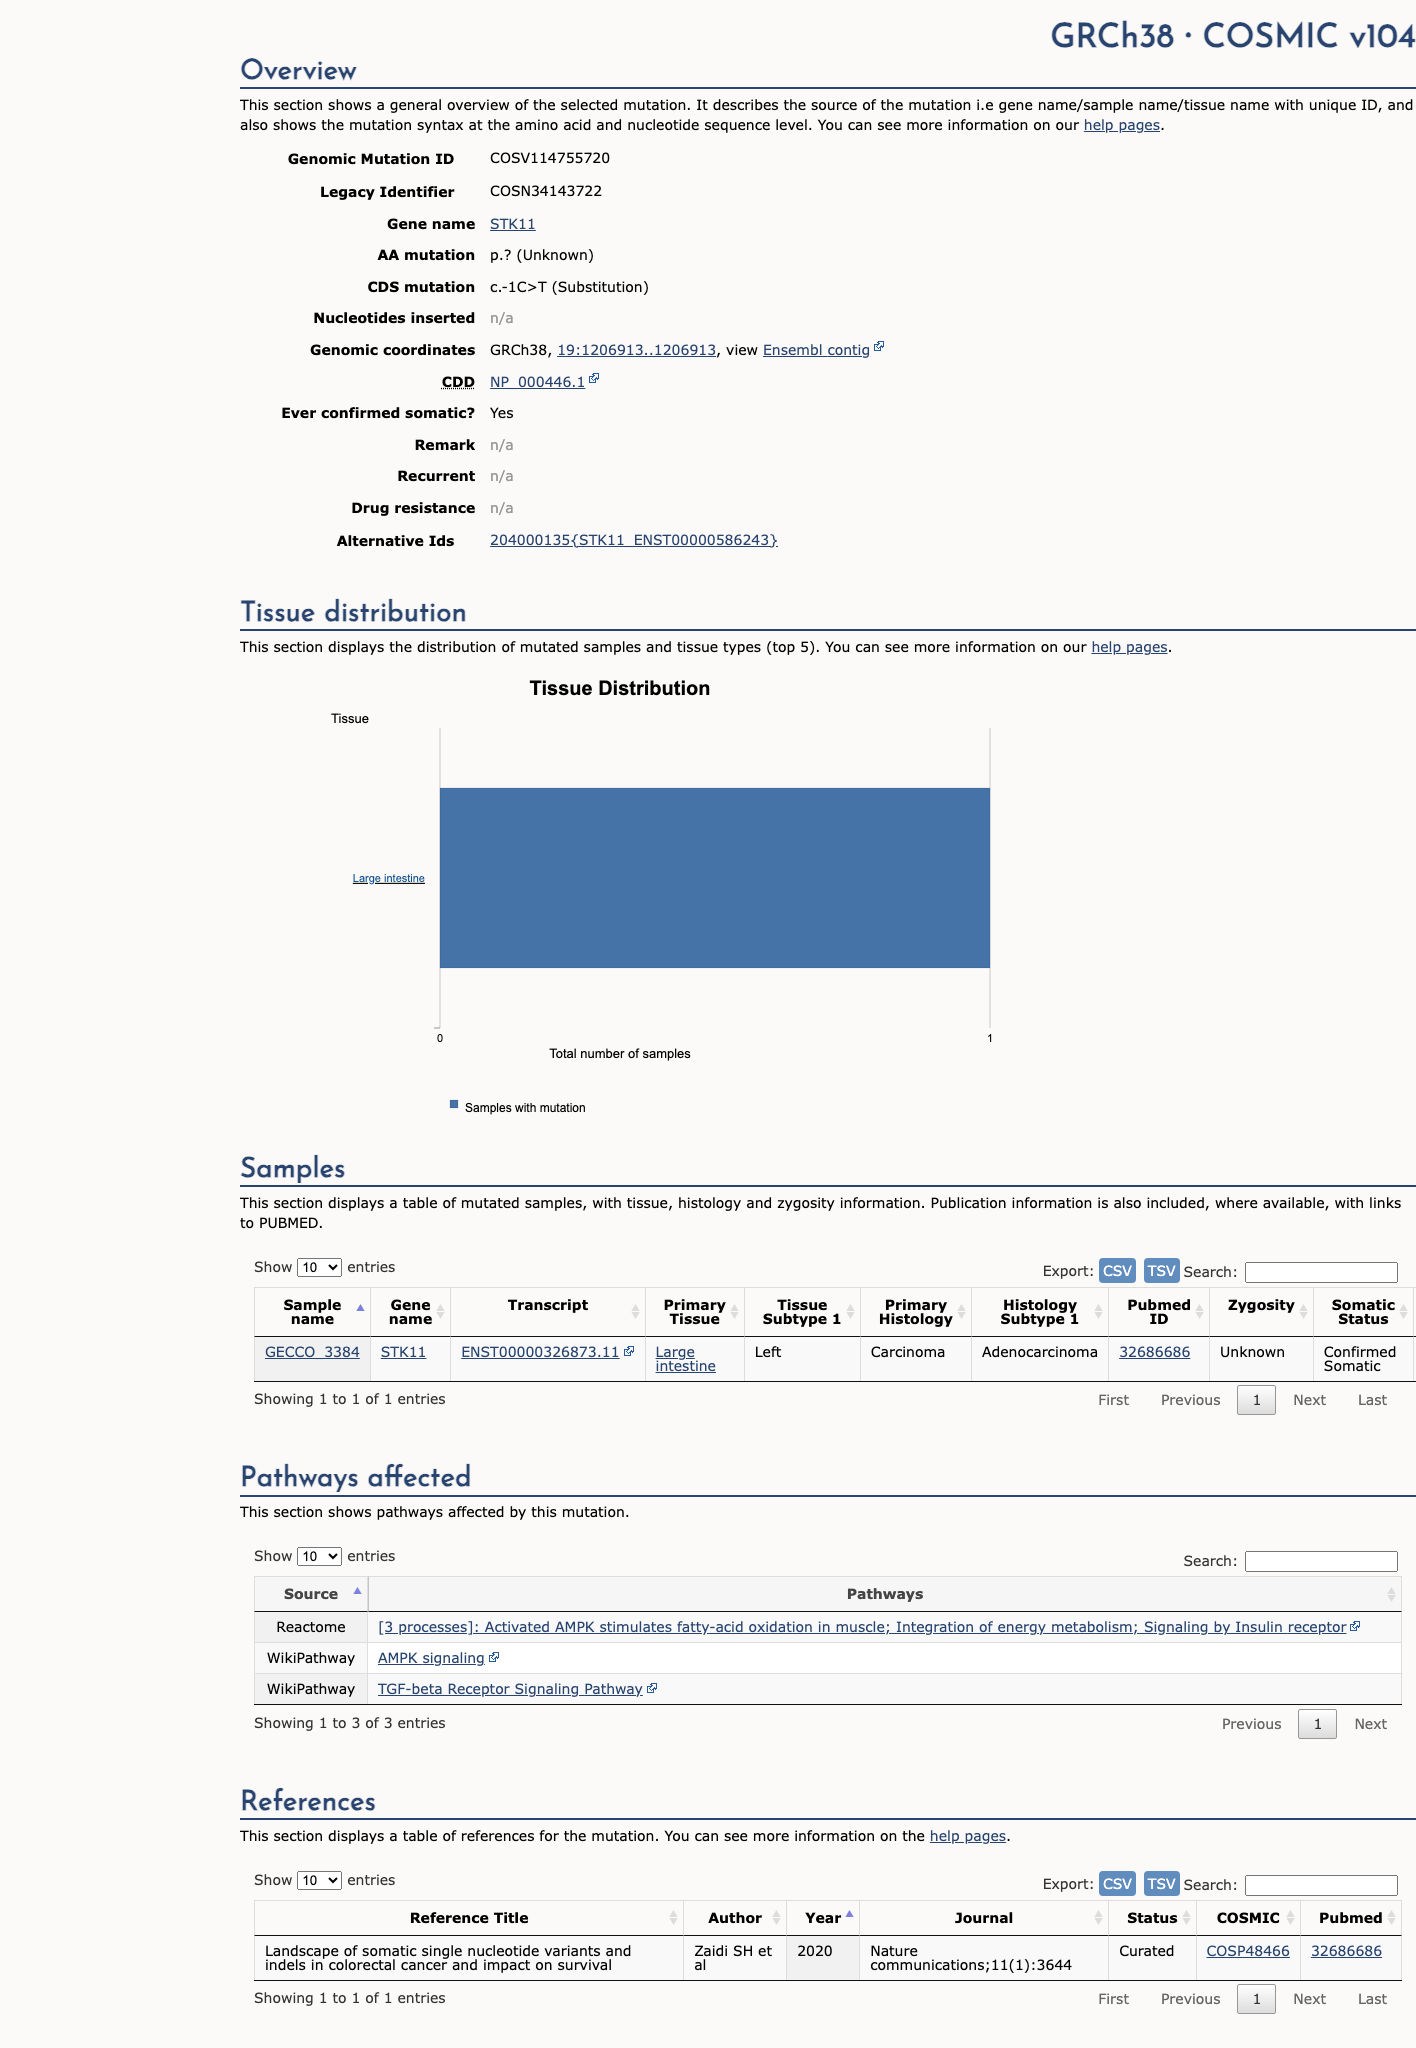
COSMIC screenshot
COSMIC
Somatic evidence
COSMIC
This variant has previously been reported in somatic cancers (COSMIC; COSV114755720, n = 1 times).
Hotspots
This variant does not lie in a statistically significant cancer hotspot.
COSMIC ↗
Literature · how each cited paper was used
2papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 6 further PMIDs triaged but not cited — see Sources & references.
Rule & framework references · cited for criterion definitions, not variant evidence
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.
26467025 ↗ A Standardized DNA Variant Scoring System for Pathogenicity Assessments in Mendelian Disorders.
Sources & reference links
7Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Triaged references · 6 PMIDs not cited in assessment
20301443 ↗ Peutz-Jeghers Syndrome. CLINVAR
23788249 ↗ ACMG recommendations for reporting of incidental findings in clinical exome and genome sequencing. CLINVAR
25980754 ↗ Identification of a Variety of Mutations in Cancer Predisposition Genes in Patients With Suspected Lynch Syndrome. CLINVAR
26389258 ↗ Cancer Genetics Risk Assessment and Counseling (PDQ®): Health Professional Versi CLINVAR
27854360 ↗ Recommendations for reporting of secondary findings in clinical exome and genome sequencing, 2016 update (ACMG SF v2.0): a policy statement of the American College of Medical Genetics and Genomics. CLINVAR
35802134 ↗ ACMG SF v3.1 list for reporting of secondary findings in clinical exome and genome sequencing: A policy statement of the American College of Medical Genetics and Genomics (ACMG). CLINVAR