Back
NM_001127510.3:c.3245C>G
p.Thr1082Ser · APC
0%
complete
Final classification
Likely Benign
BS1BP1
APC
c.3245C>G
p.Thr1082Ser
missense · exon 17

APC is a tumor suppressor gene whose protein acts as a key brake on the Wnt signaling pathway by helping mark the growth-promoting protein beta-catenin for destruction; it also participates in cell migration, adhesion, and apoptosis. Inherited mutations in APC cause familial adenomatous polyposis (FAP), an autosomal dominant condition characterized by numerous colorectal polyps and a very high risk of colorectal cancer, with related forms including attenuated FAP and gastric adenocarcinoma and proximal polyposis of the stomach (GAPPS). Somatic APC mutations are found in roughly 50-80% of sporadic colorectal cancers, where they act as early tumor-initiating events, and are also observed in some breast, stomach, and prostate cancers. Loss of APC function leaves Wnt signaling abnormally active, driving uncontrolled cell growth, which makes APC a central tumor suppressor in colorectal cancer.

This variant

APC is an autosomal dominant tumor suppressor gene in which truncating loss-of-function variants cause familial adenomatous polyposis and a high colorectal cancer risk, so a missense change such as p.Thr1082Ser is interpreted against the gene-specific InSiGHT APC VCEP framework rather than generic ACMG rules.

Transcript
NM_001127510.3
HGVS · transcript:coding
NM_001127510.3:c.3245C>G
GRCh38
chr5:112838839 C>G
GRCh37
chr5:112174536 C>G
Likely Benign: BS1 (strong, gnomAD non-cancer popmax filtering AF 0.0039% exceeds the 0.001% threshold) satisfies the APC VCEP's Rule26 Benign-Strong combination.
Classification rationale
BS1BP1 Likely Benign
APC c.3245C>G missense · exon 17

Likely Benign driver: BS1 (strong) - gnomAD v2.1.1 non-cancer popmax filtering AF 0.0039% exceeds the APC VCEP threshold of 0.001%. Likely Benign: BP1 (supporting) - missense change at residue 1082 sits outside the VCEP-excepted beta-catenin repeat codons 1021-1035. Likely Benign: APC VCEP Rule26 is met by one Benign-Strong criterion, while no pathogenic combination rule is satisfied.

BS1 + BP1 → Likely Benign
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_001127510.3 · variants mapped to exon structure
APC NM_001127510.3
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 15 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
BS1 strong Benign
Met (strong): popmax filtering AF 0.0039% in gnomAD v2.1 non-cancer exceeds the VCEP BS1 threshold of 0.001%.
Governing specification: InSiGHT APC VCEP v2.1. BS1 strength rule: 'GnomAD Popmax Filtering Allele Frequency (AF) >= 0.001% (0.00001).' instructionsToUse: 'General recommendation: Use the non-cancer dataset from gnomAD (v2.1.1)'.gnomAD v2.1.1 non-cancer subset: grpmax filtering AF 3.865e-05 (0.003865%); exome AC 11, AN 236026, exome AF 4.6605035e-05; highest subpopulation European (Finnish) AF 0.00011951238945103975 (0.011950%), above the 0.001% threshold.gnomAD v2.1 all-comers: grpmax filtering AF 3.442e-05 (0.003442%); gnomAD v4.1 all-comers: grpmax filtering AF 7.076e-05 (0.007076%). Both exceed 0.001%.
BP1 supporting Benign
Met (supporting): p.Thr1082Ser is a missense change at residue 1082, outside the beta-catenin repeat codons 1021-1035 that the APC VCEP excepts.
ClinGen InSiGHT APC VCEP specification v2.1 (cspec), BP1 Supporting rule: 'BP1 is applicable to APC with the exception of missense variants located in the first 15-amino acid repeat of the beta-catenin binding domain (codon 1021-1035).'The variant under assessment is a missense change at residue 1082 (p.Thr1082Ser), which is outside codons 1021-1035, so the VCEP exception does not remove BP1 applicability.APC is a gene in which primarily truncating (loss-of-function) variants are known to cause familial adenomatous polyposis 1 (MONDO:0021056); this is the mechanism premise of BP1.
Assessed · not applied · 9 not met · 6 not assessed
Pathogenic
PS1 Not met: no established pathogenic variant carries the same p.Thr1082Ser change; APC's only Likely Pathogenic missense variants are p.Asn1026Ser and p.Ser1028Arg.
PS2 Not assessed: no proband with confirmed parentage or documented de novo occurrence, so no de novo score could be counted against the VCEP >=4/2-3.5/1-1.5 thresholds.
PS3 Not assessed: no RNA or protein functional assay has been reported for APC p.(Thr1082Ser), so the APC VCEP PS3 requirements cannot be evaluated.
PS4 Not met: 0 phenotype points assignable (no polyposis phenotype documented) against the >=1 phenotype point required for PS4_Supporting.
PM2 Not met: non-cancer exome AF 0.00466% (11 alleles) exceeds the VCEP PM2 threshold of 0.0003% for allele count greater than 1.
PM5 Not met: residue 1082 has no pathogenic alternate missense; the APC VCEP restricts PM5 to residues 1026 and 1028.
PM6 Not assessed: no de novo occurrence, even with unconfirmed parentage, is reported, so no VCEP PM6 de novo score (0.5-3.5) can be assigned.
PP1 Not assessed: no pedigree or meiosis data are reported, so co-segregation cannot be counted against the VCEP >=7/5-6/3-4 meiosis thresholds.
PP3 Not met: for APC missense variants PP3 uses splice predictors, and SpliceAI max delta 0.001 is far below the 0.2 supporting cutoff.
Benign
BA1 Not met: highest popmax filtering AF 0.0039% (gnomAD v2.1 non-cancer) versus the VCEP BA1 stand-alone threshold of 0.1%.
BS2 Not met: zero homozygotes across every gnomAD dataset, and the VCEP bars counting gnomAD heterozygous carriers as healthy individuals.
BS3 Not assessed: no protein assay showing retained beta-catenin-regulated transcription has been reported for APC p.(Thr1082Ser), so the APC VCEP BS3 protein-assay branch is unevaluable.
BS4 Not assessed: no genotyped affected relative without the variant is reported, so no phenotype-point deficit can be scored for VCEP BS4.
BP2 Not met: no source reports c.3245C>G in trans with a pathogenic APC variant, and none of the required >=3 unknown-phase co-occurrences exist.
BP5 Not met: no (likely) pathogenic variant in another adenomatous polyposis gene is documented, so the alternate-molecular-basis condition for BP5_Supporting is absent.
N/A · 11 PVS1 · PM1 · PM3 · PM4 · PP2 · PP4 · PP5 · BP3 · BP4 · BP6 · BP7
Research & evidence
Population frequency · supports benign
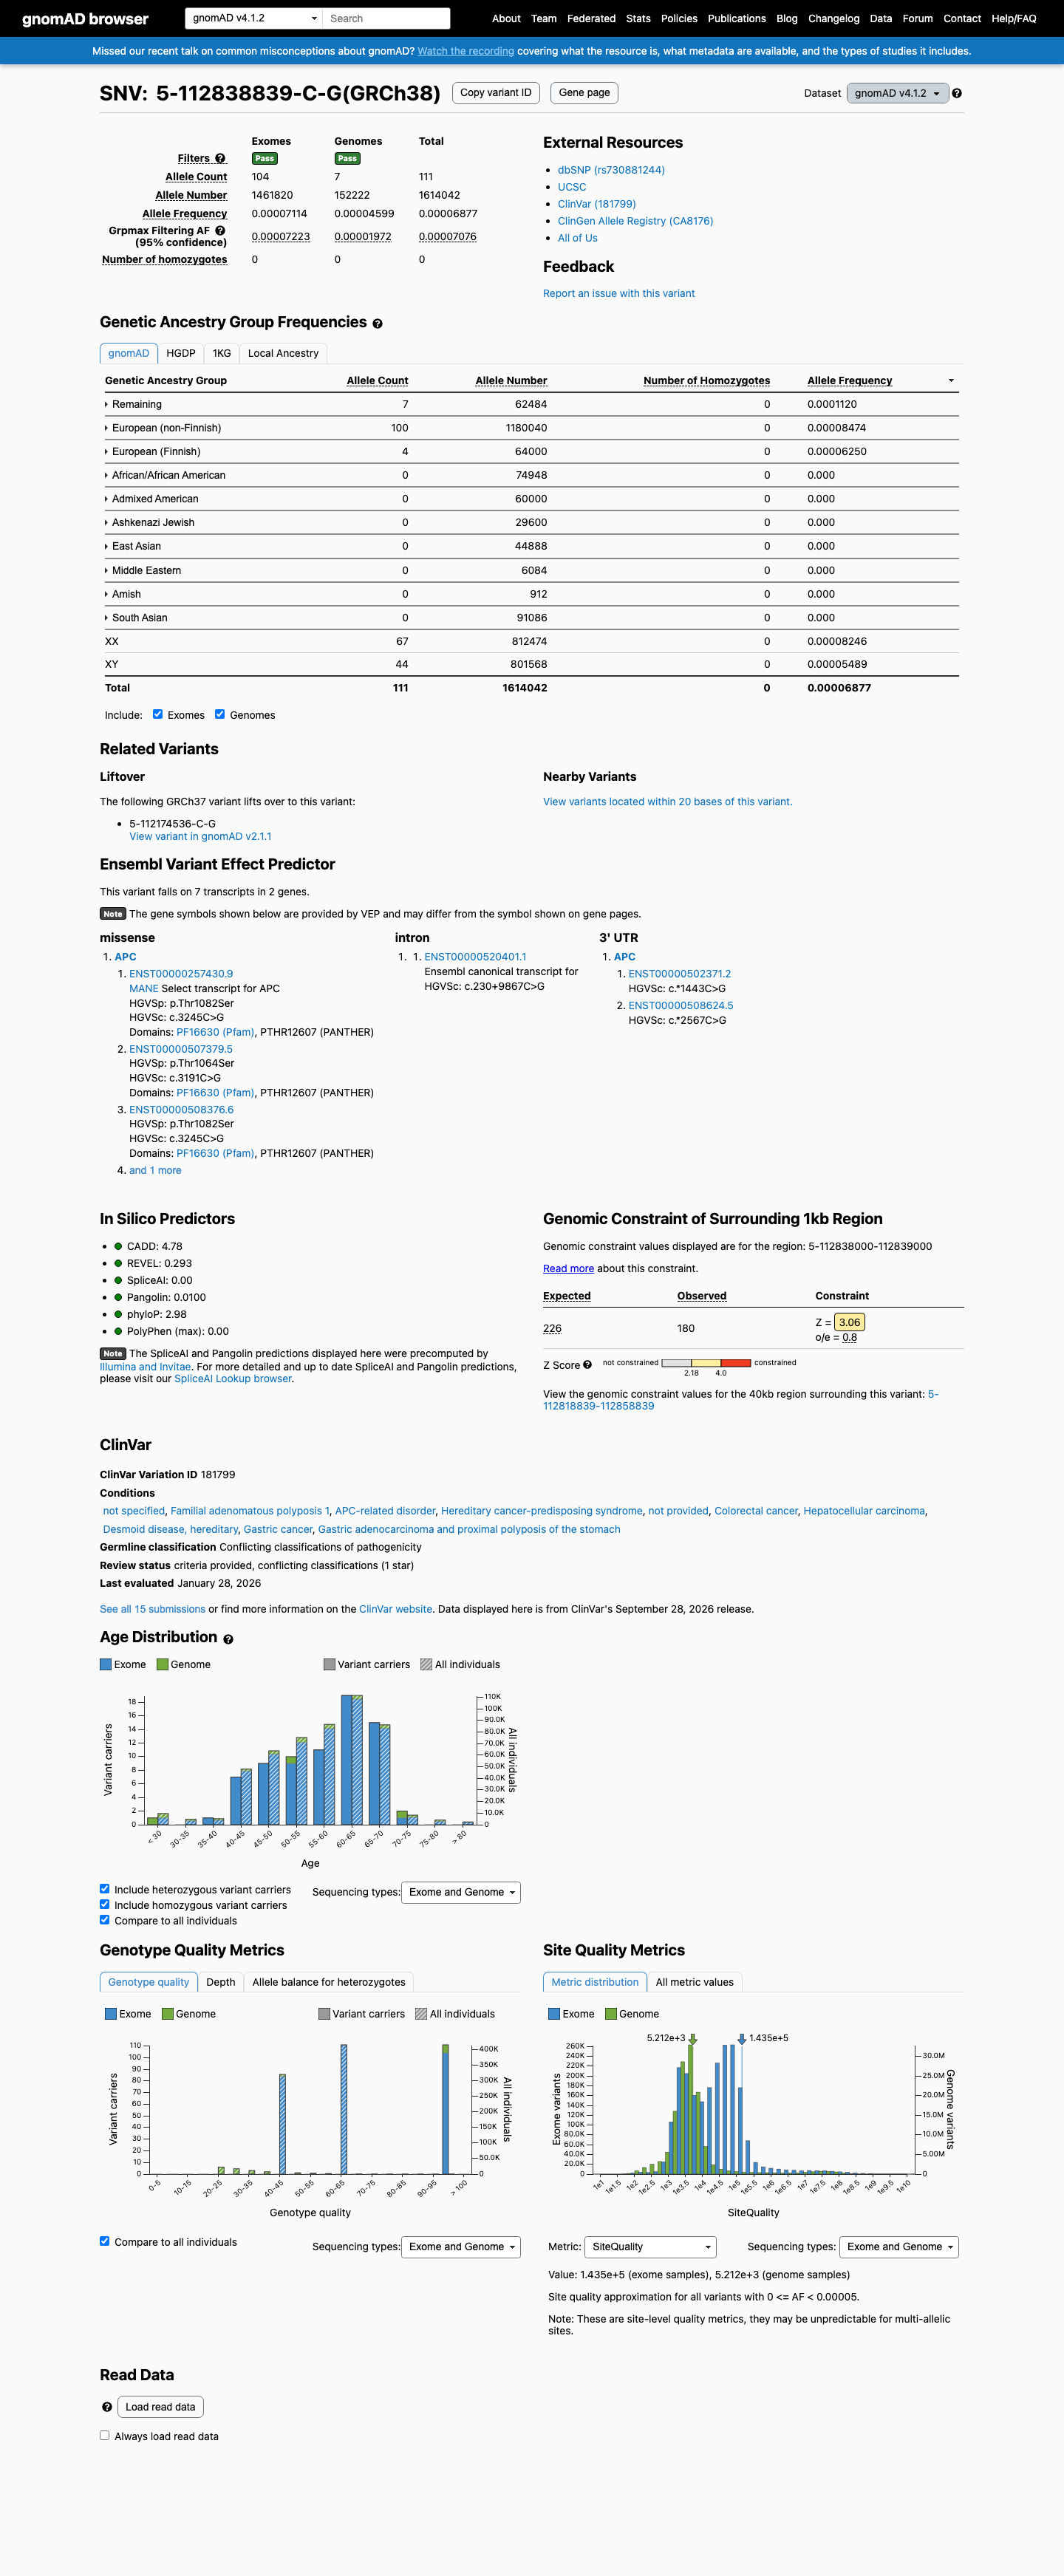
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
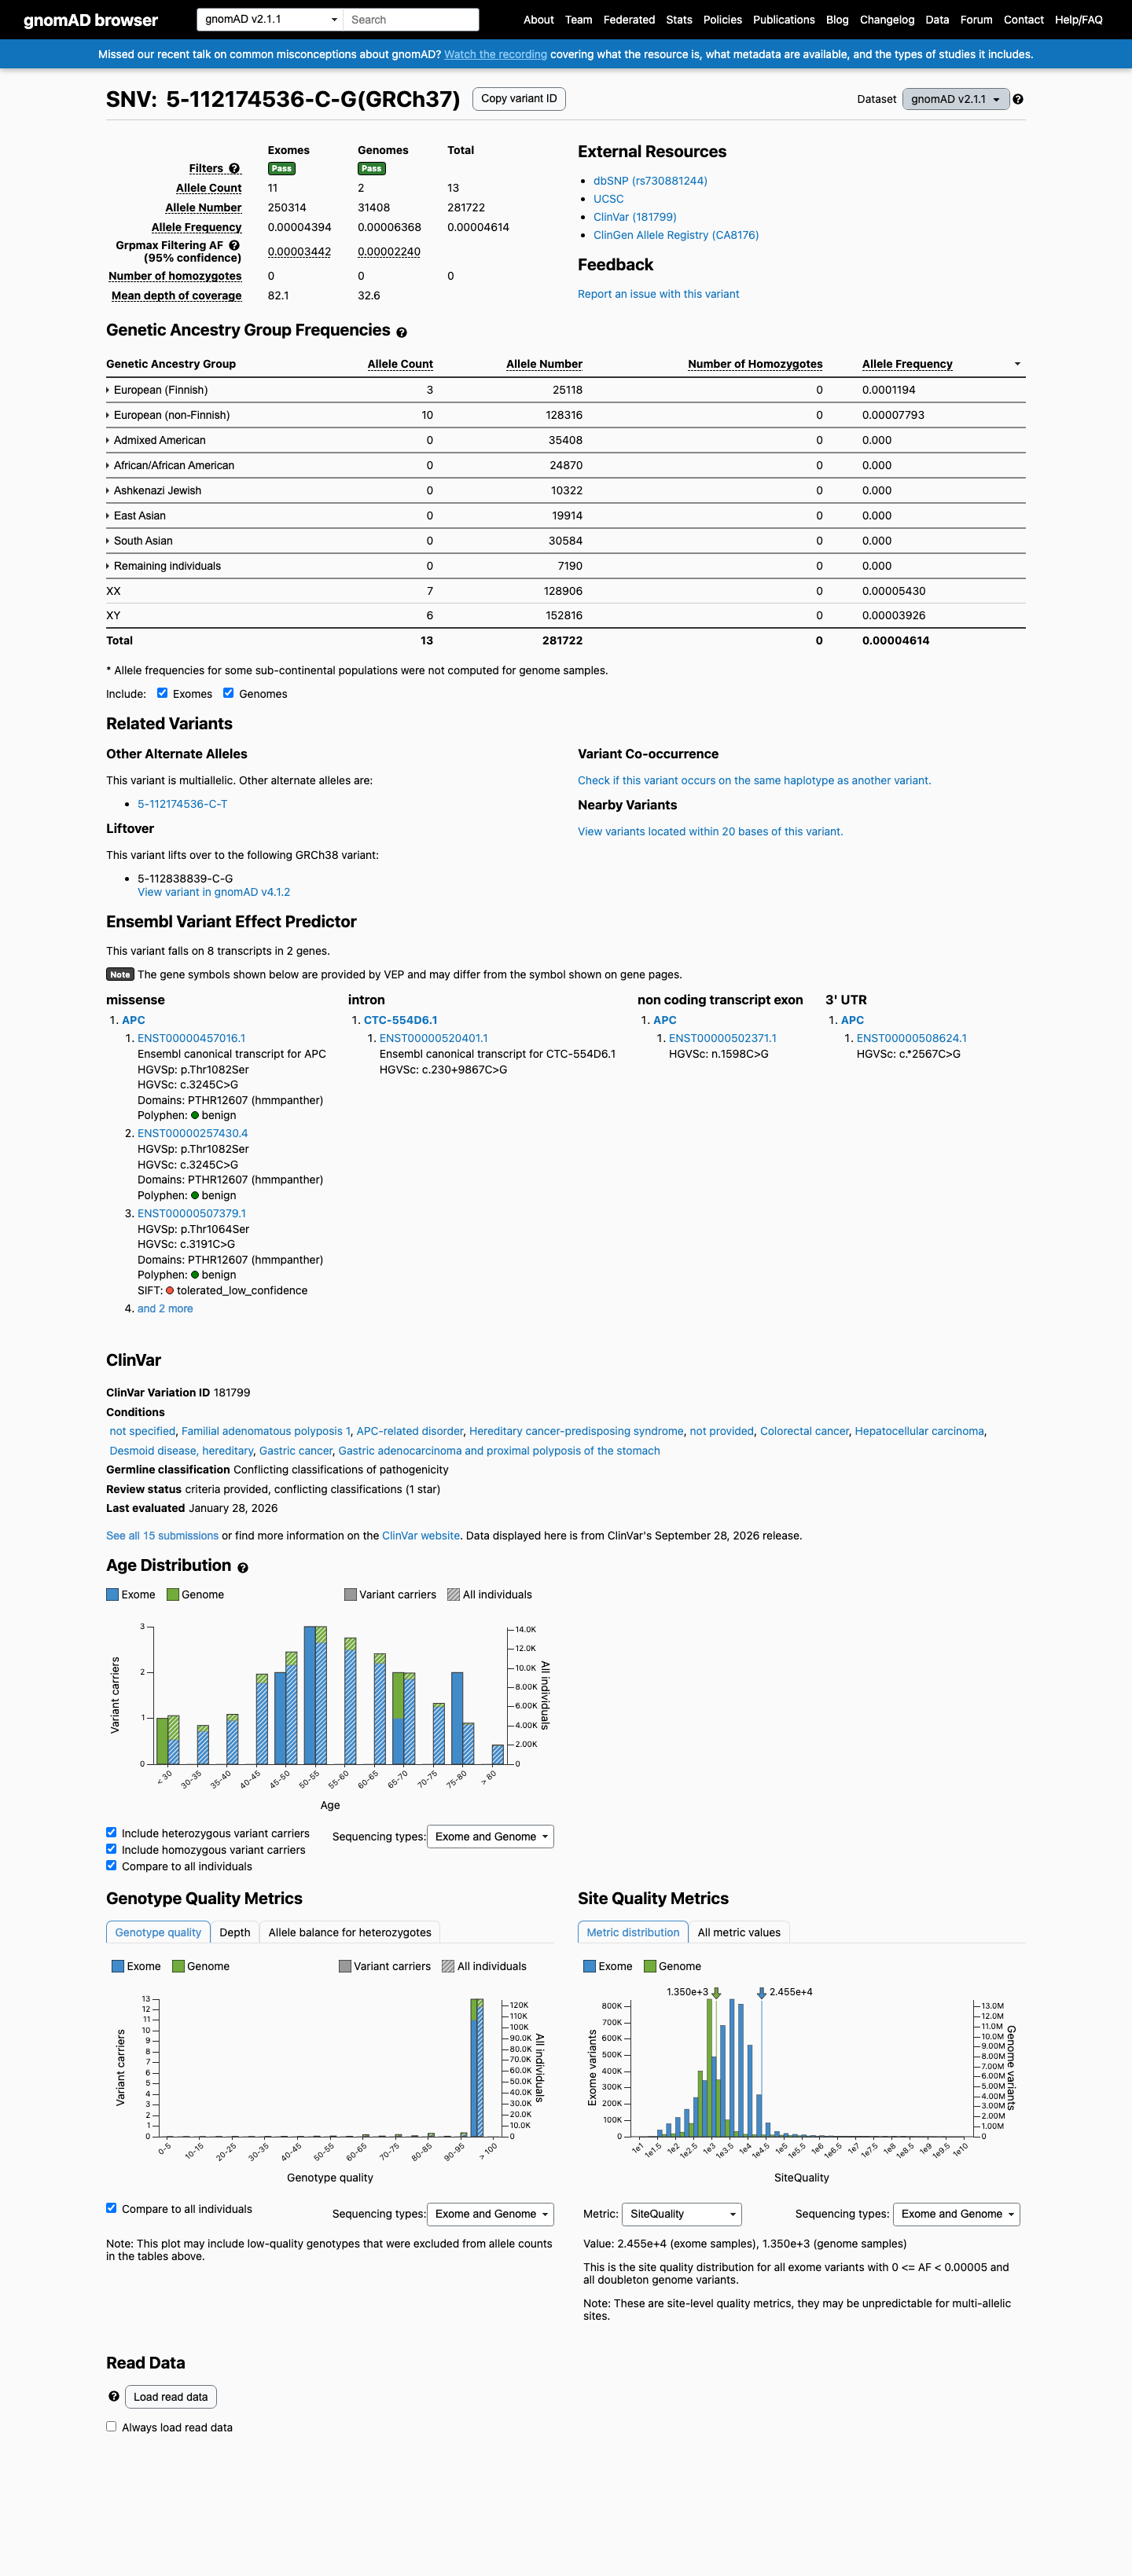
This variant is present in gnomAD v4.1 (AF= 6.87714e-05; MAF= 0.00688%, 111/1614042 alleles, homozygotes = 0) and has highest observed frequency in the Remaining individuals population (AF= 0.000112029; MAF= 0.01120%, 7/62484 alleles, homozygotes = 0); grpmax FAF= 7.076e-05.
v2.1
This variant is present in gnomAD v2.1 (AF= 4.61448e-05; MAF= 0.00461%, 13/281722 alleles, homozygotes = 0) and has highest observed frequency in the European (Finnish) population (AF= 0.000119436; MAF= 0.01194%, 3/25118 alleles, homozygotes = 0); grpmax FAF= 3.442e-05.
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.0069% · 111 / 1,614,042
0 hom · FAF 0.0071%
Remaining individuals
7 / 62,484
0.011%
European (non-Finnish)
100 / 1,180,040
0.0085%
European (Finnish)
4 / 64,000
0.0063%
+ 7 not observed (Admixed American, Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American)
gnomAD v2.1
0.0046% · 13 / 281,722
0 hom · FAF 0.0034%
European (Finnish)
3 / 25,118
0.012%
European (non-Finnish)
10 / 128,316
0.0078%
+ 6 not observed (African/African American, Admixed American, Ashkenazi Jewish, East Asian, Remaining individuals, South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
ClinVar screenshot
ClinVar
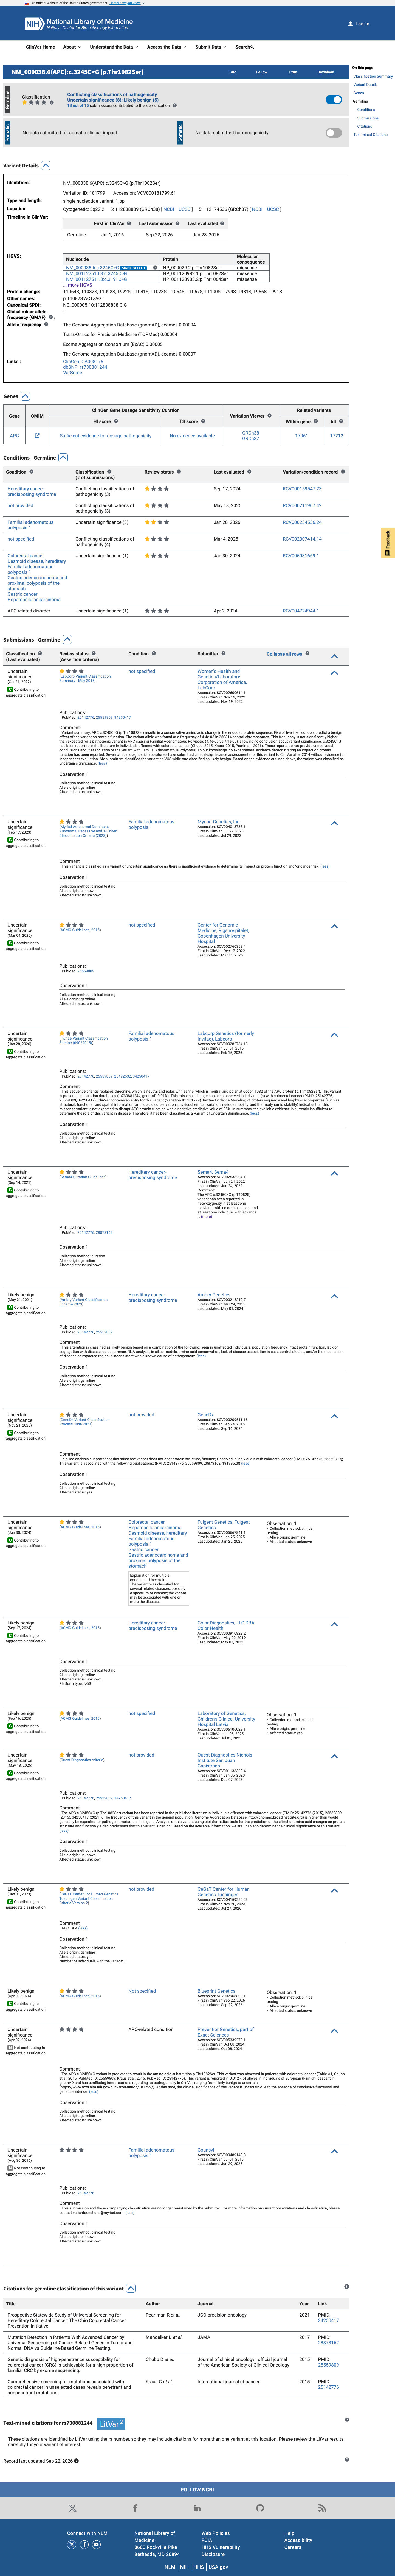
This variant has been reported in ClinVar as Uncertain significance (9 clinical laboratories) and as Likely benign (5 clinical laboratories). (ClinVarID = 181799)
SpliceAI screenshot
In silico
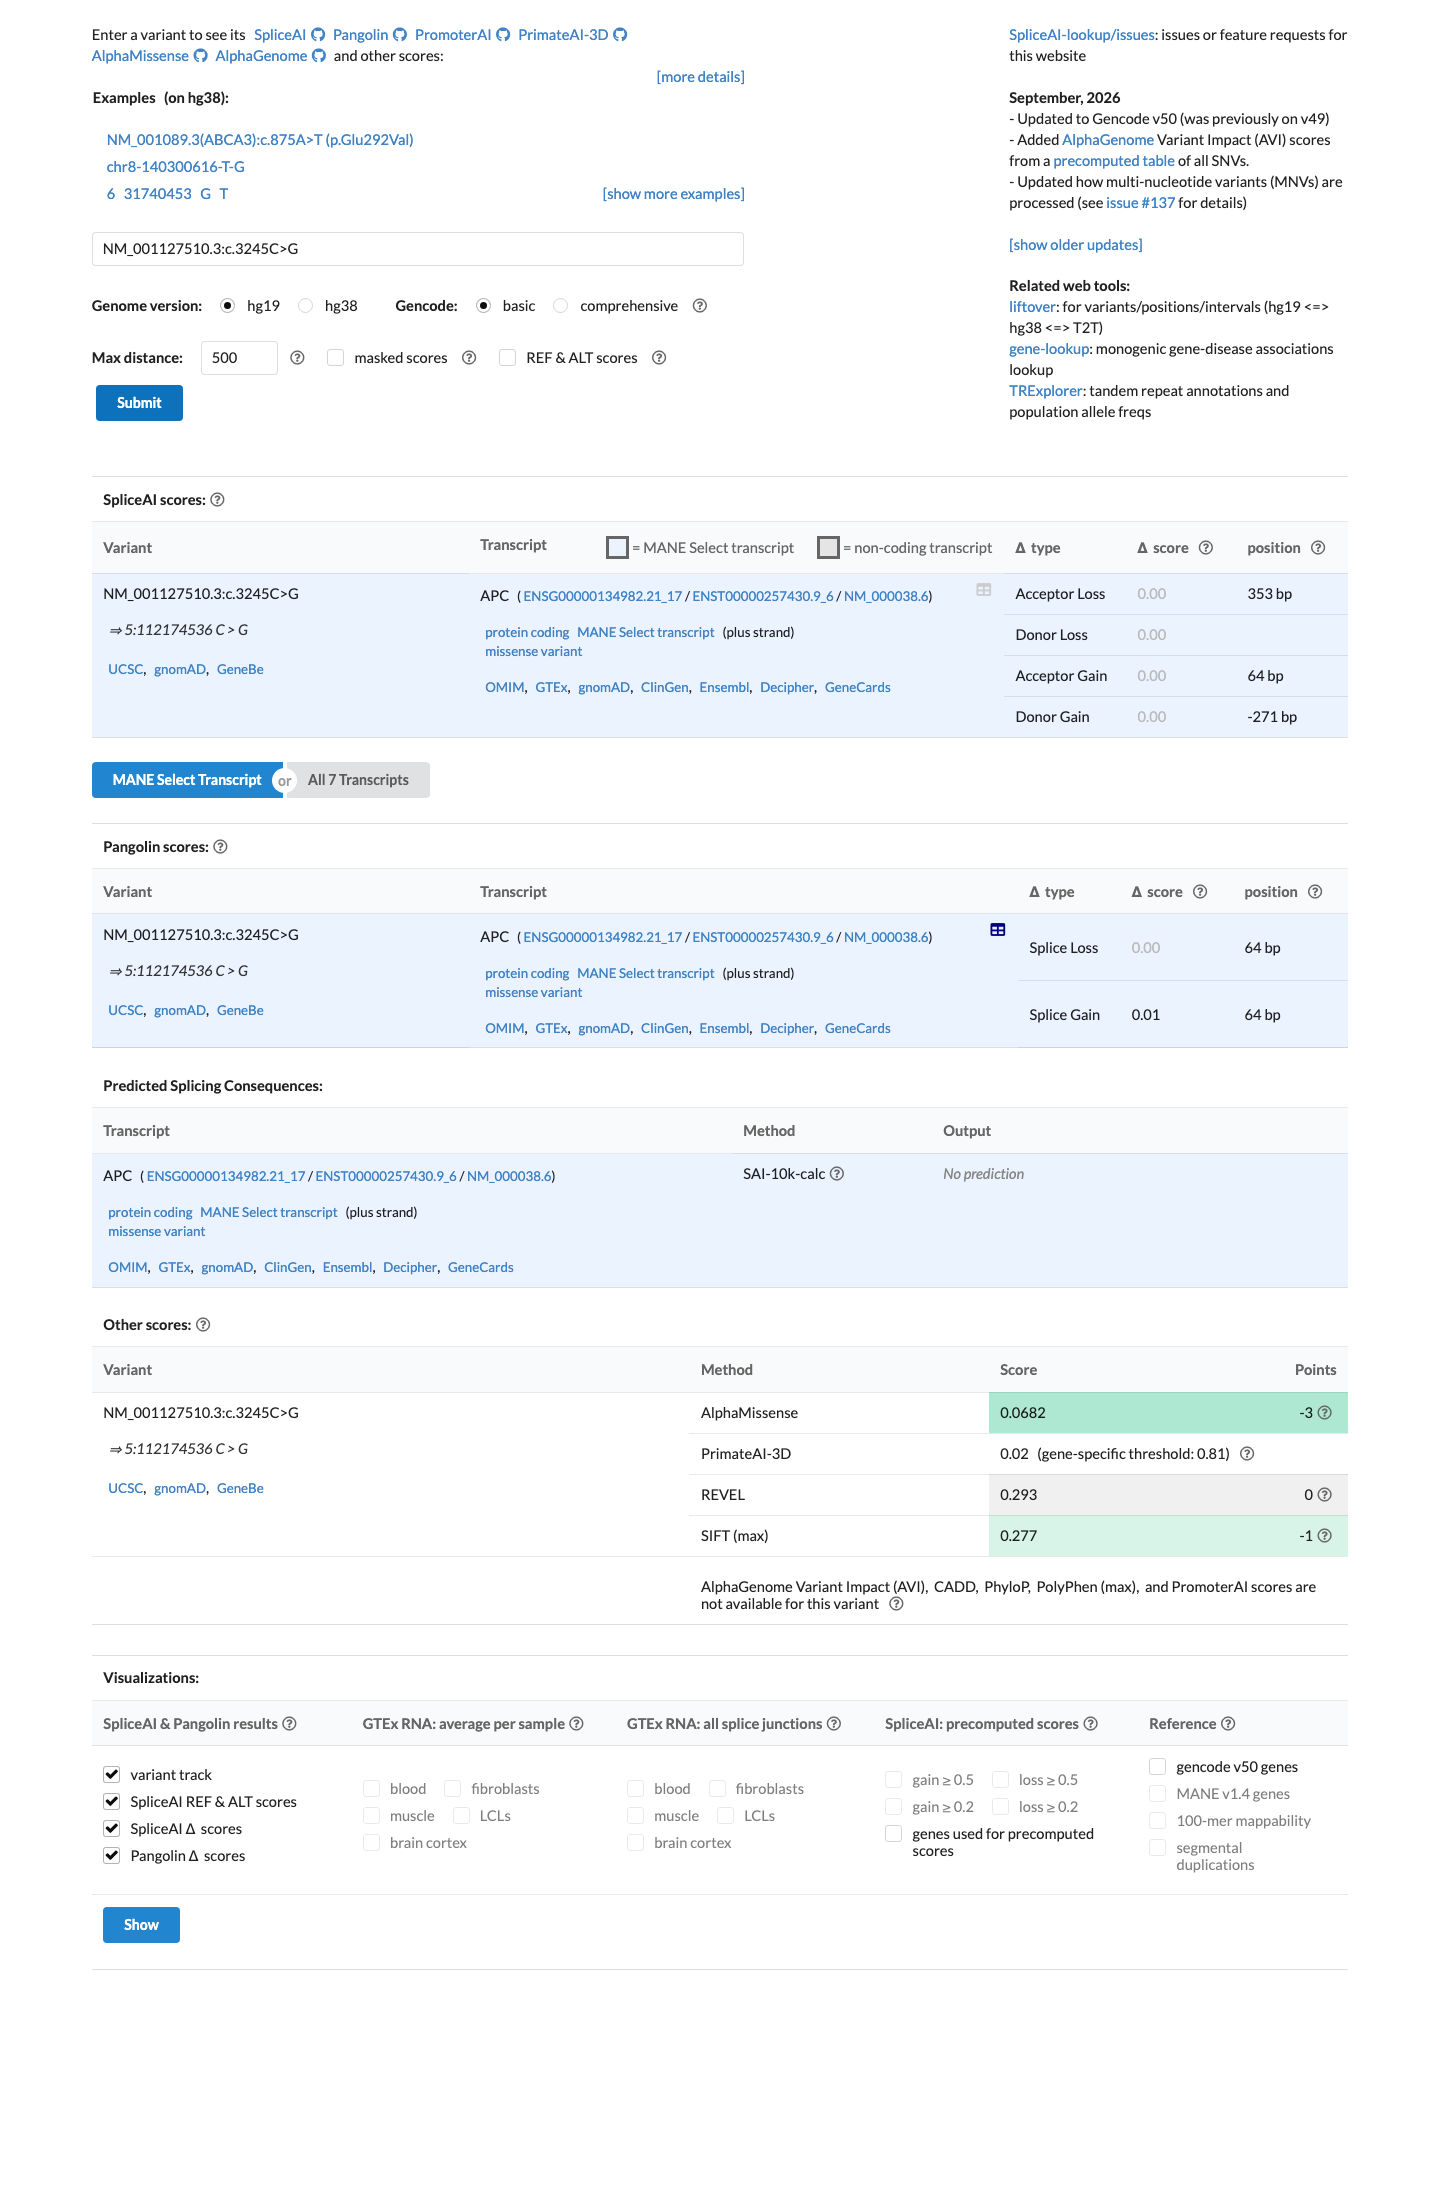
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.00). REVEL score = 0.293. BayesDel score = -0.278933.
Functional / OncoKB screenshot
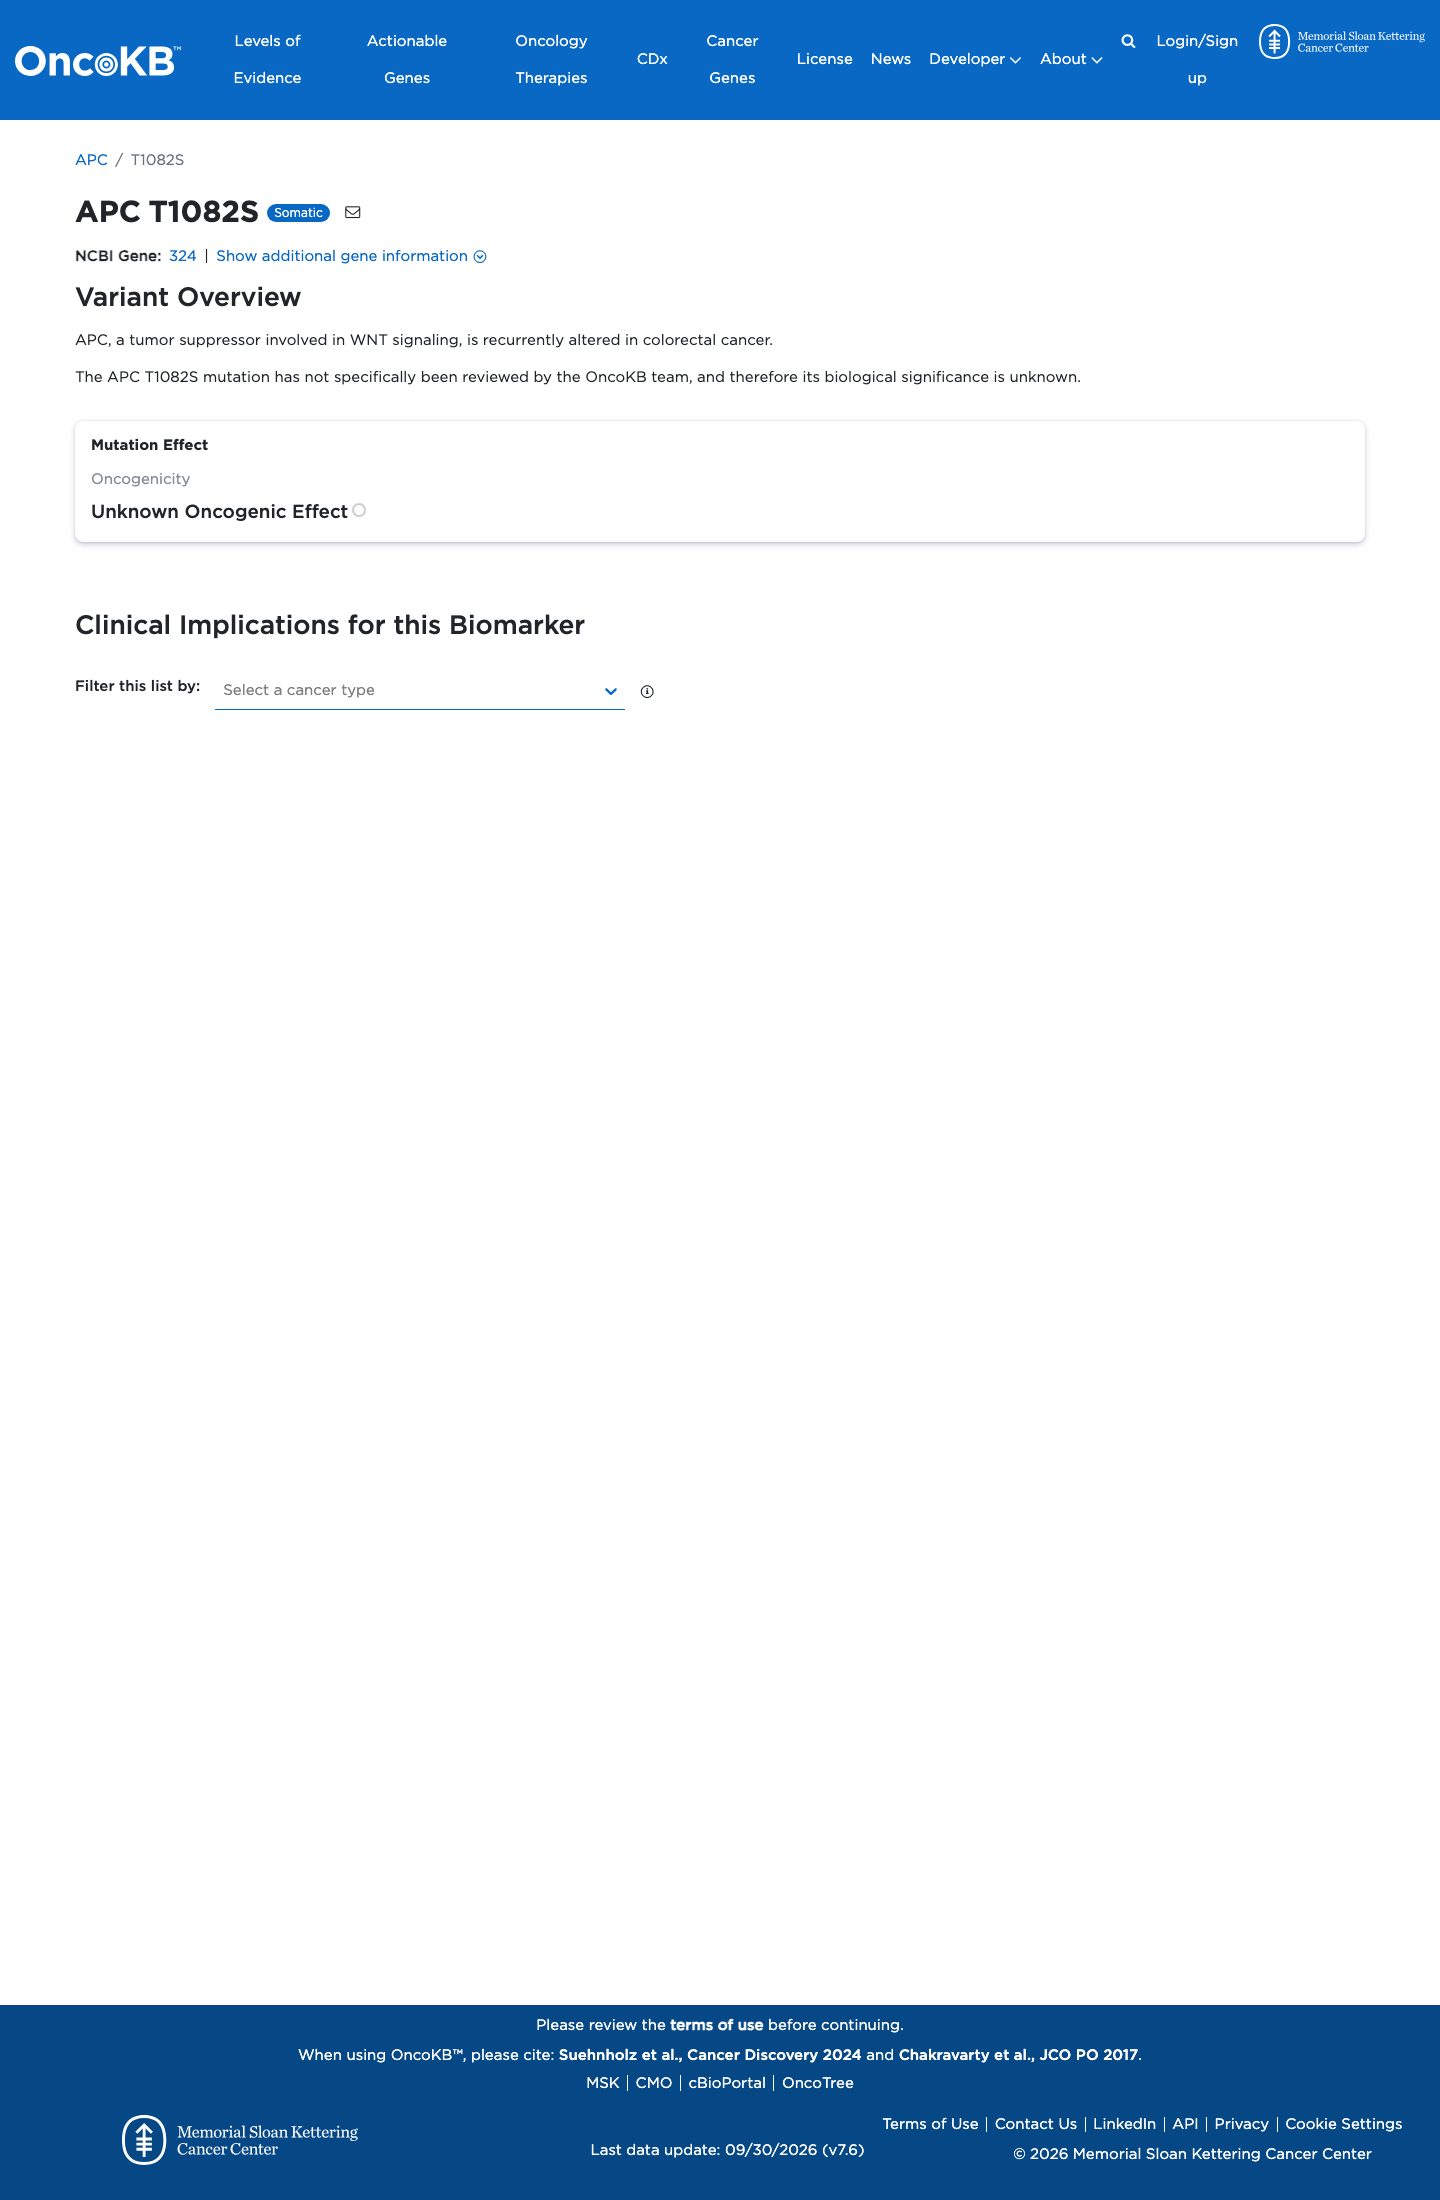
Functional Unknown Oncogenic Effect
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. APC, a tumor suppressor involved in WNT signaling, is recurrently altered in colorectal cancer.
OncoKB ↗
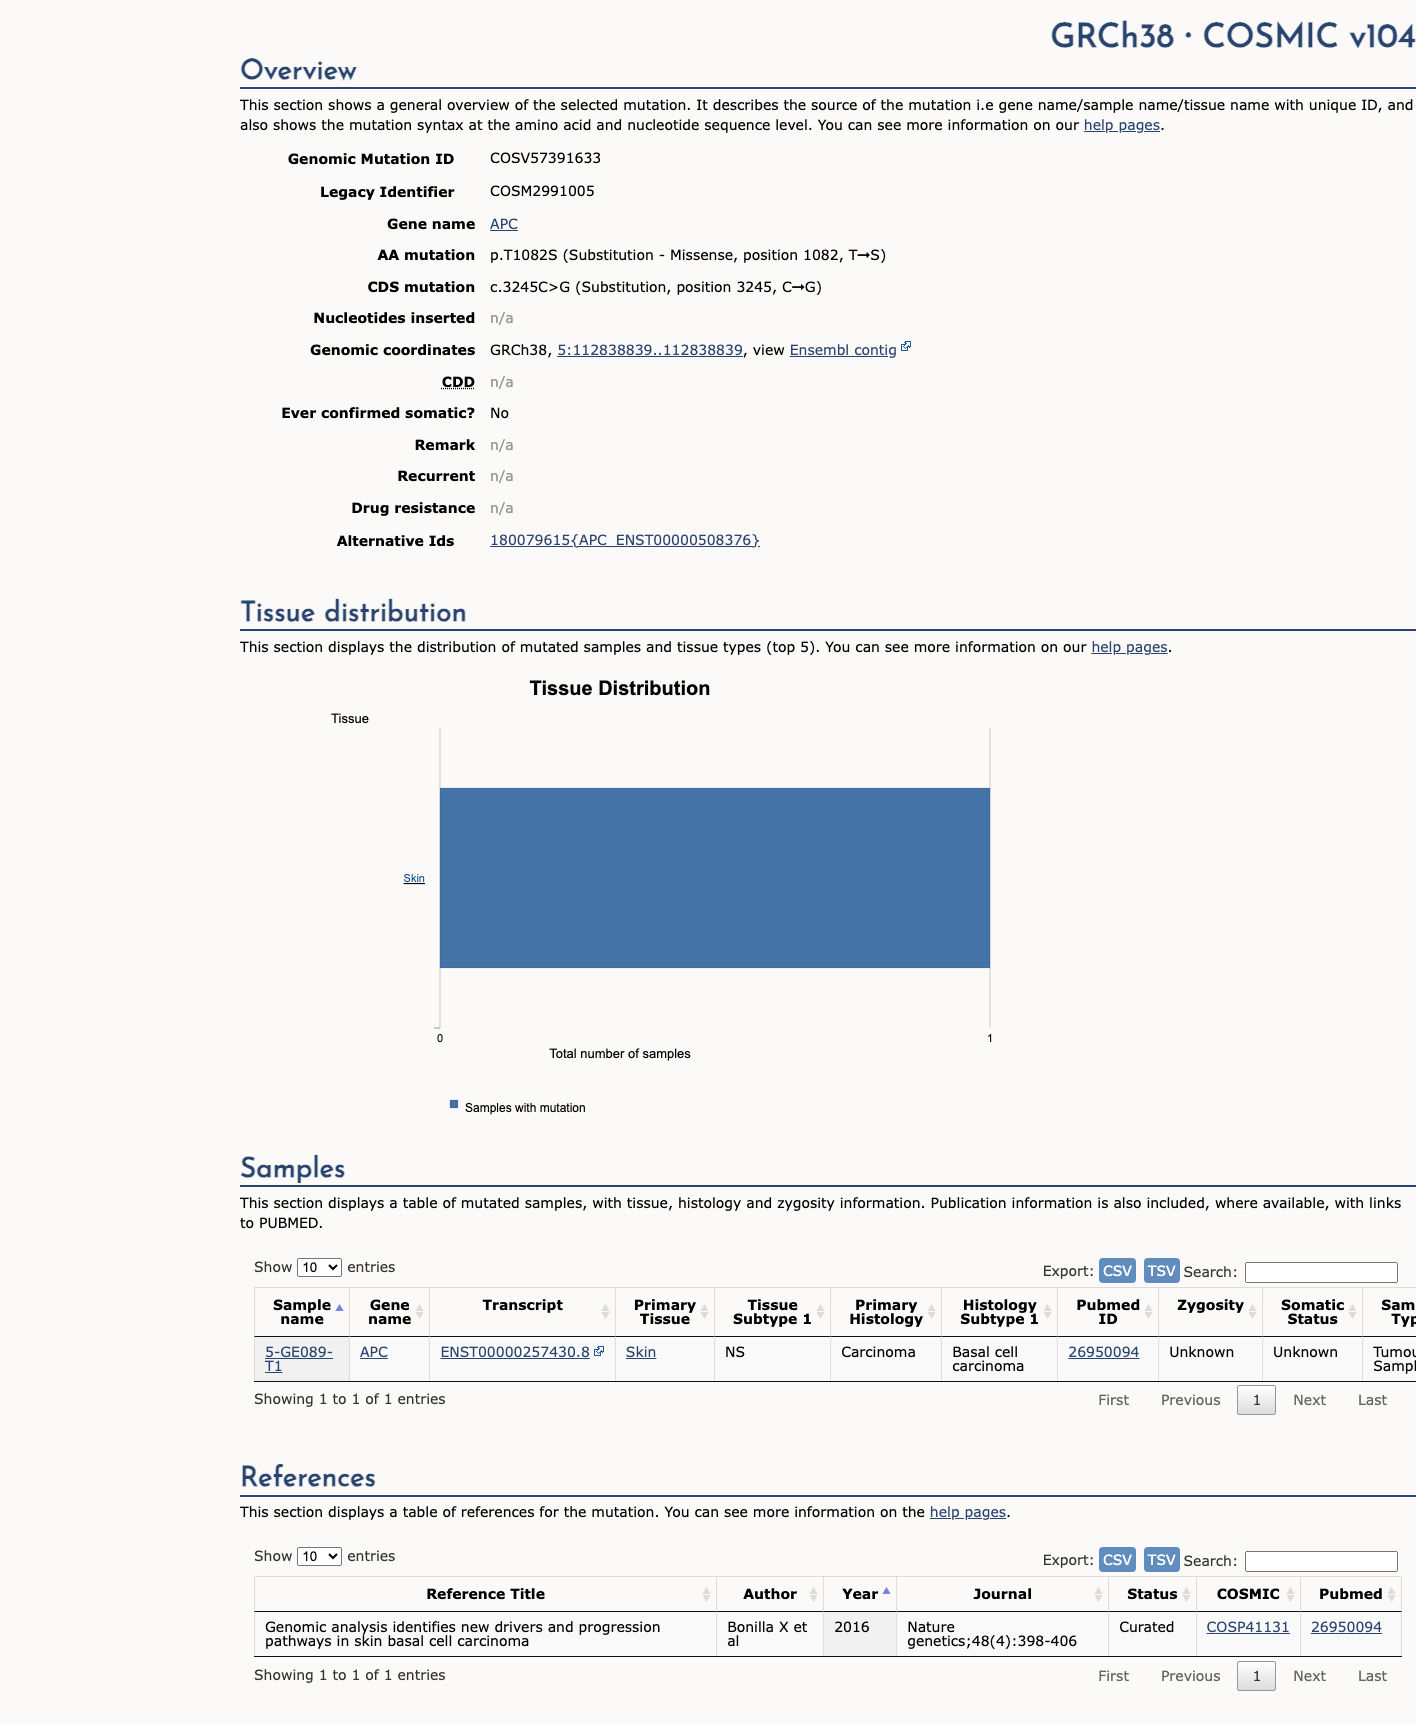
COSMIC screenshot
COSMIC
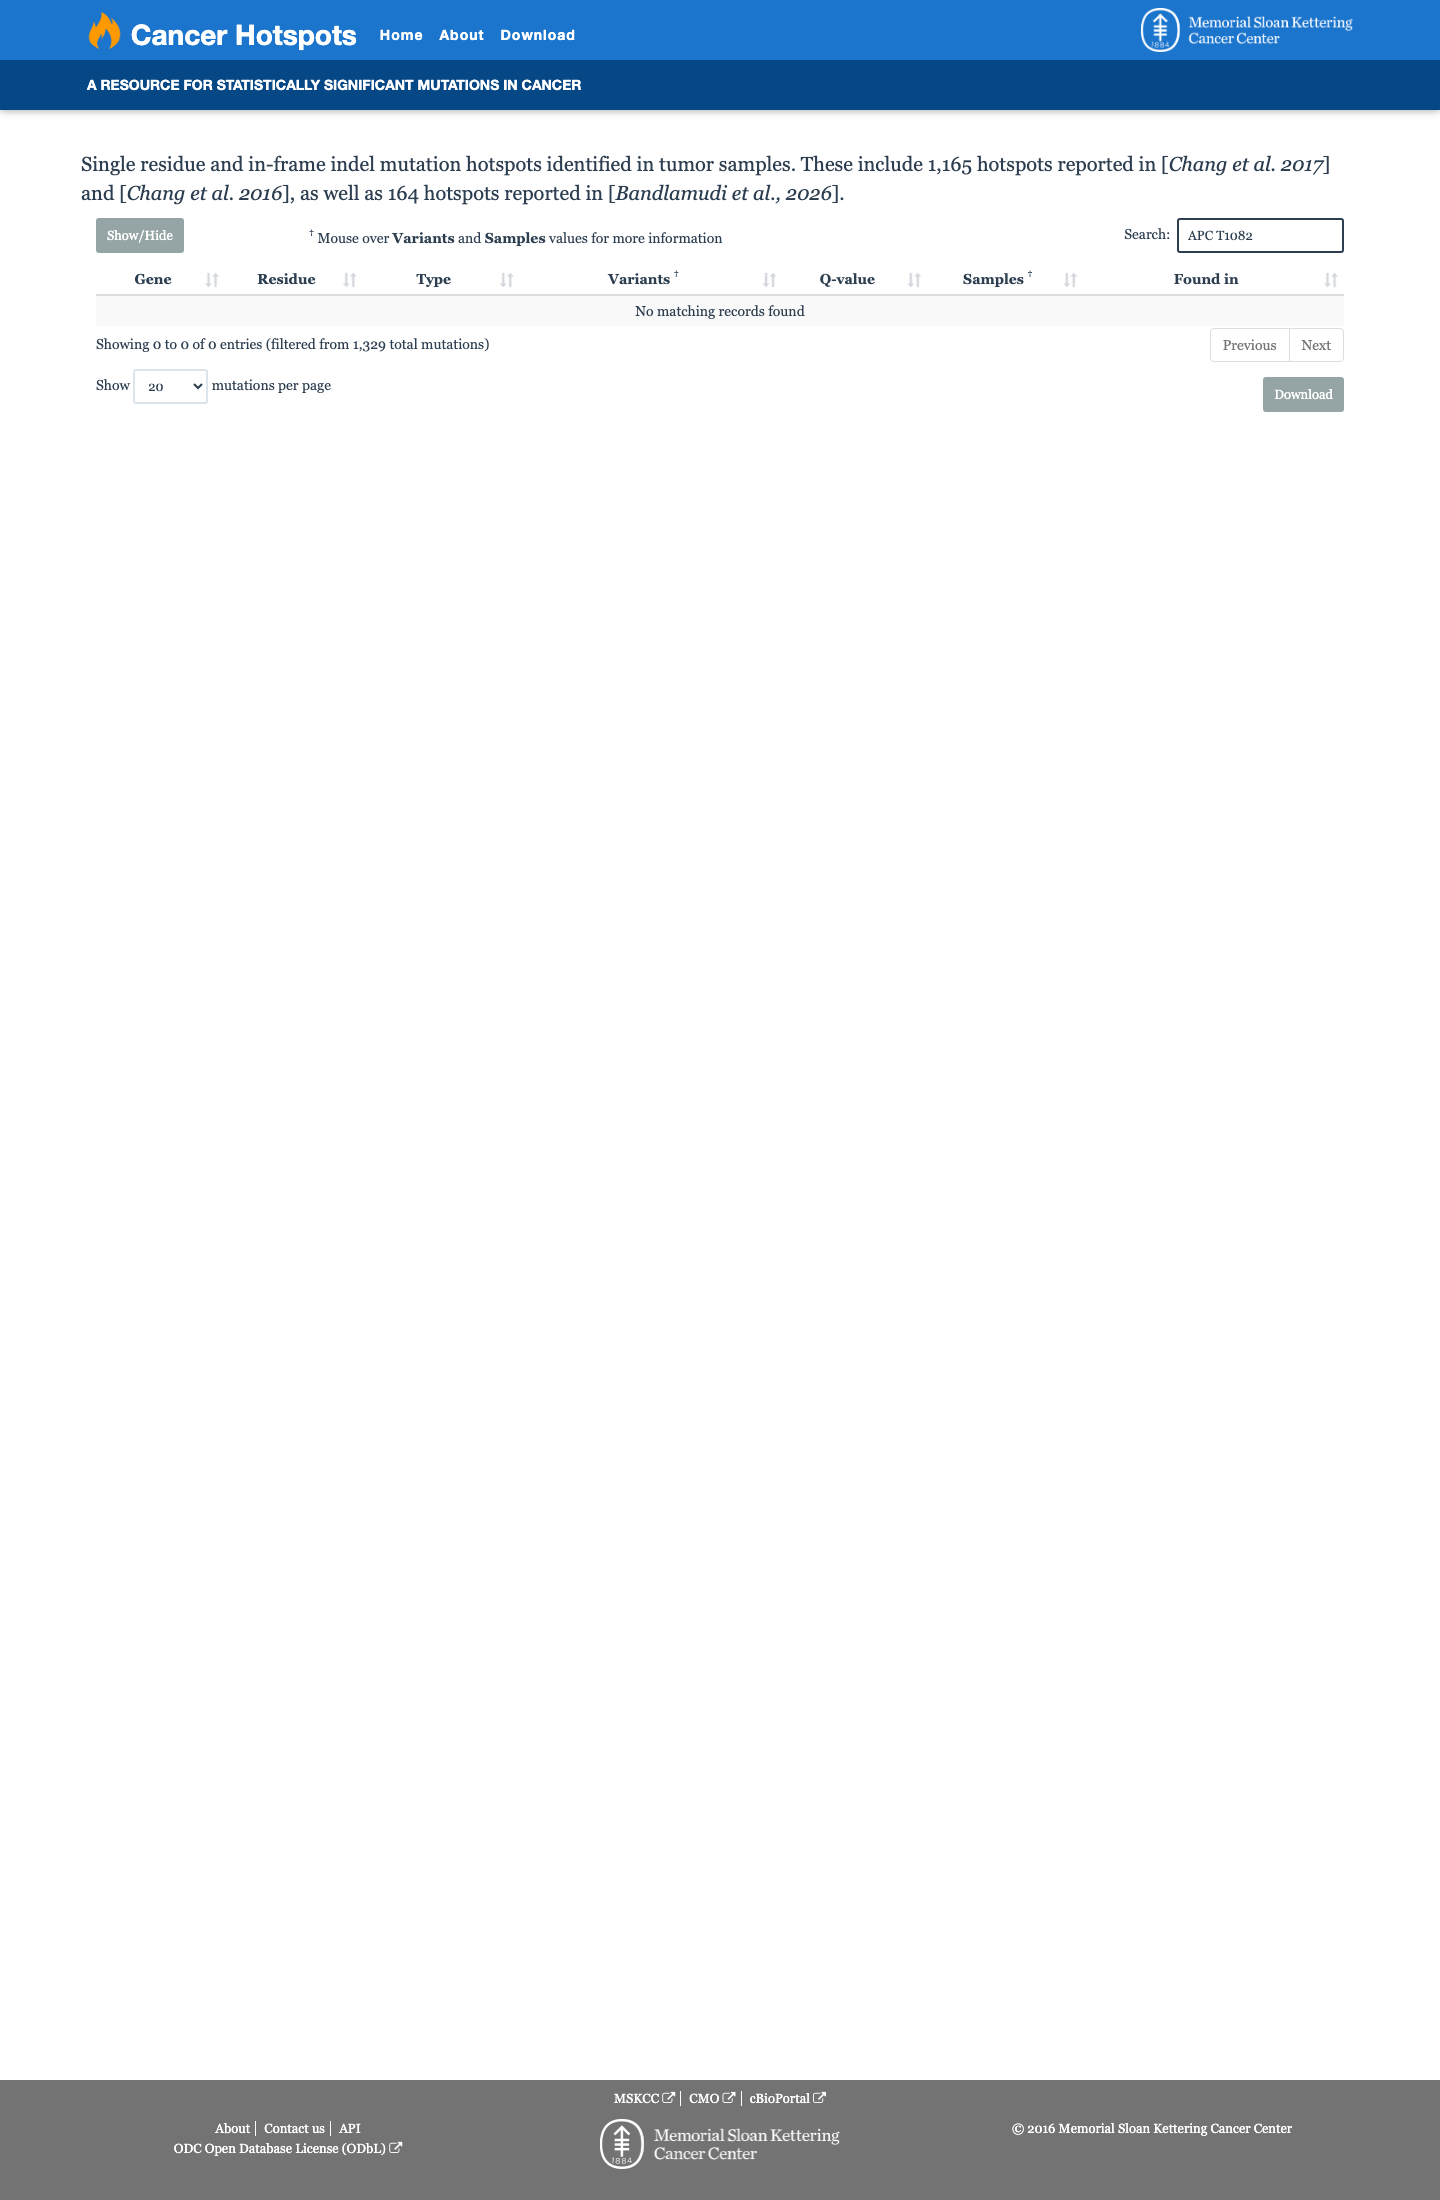
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has previously been reported in somatic cancers (COSMIC; COSV57391633, n = 1 times).
Hotspots
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
1papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 7 further PMIDs triaged but not cited — see Sources & references.
Rule & framework references · cited for criterion definitions, not variant evidence
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint co
Sources & reference links
9Sources
CSpec VCEP
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 7 PMIDs not cited in assessment
25142776 ↗ Comprehensive screening for mutations associated with colorectal cancer in unselected cases reveals penetrant and nonpenetrant mutations. CLINVAR
25452455 ↗ Hereditary colorectal cancer syndromes: American Society of Clinical Oncology Cl CLINVAR
25559809 ↗ Genetic diagnosis of high-penetrance susceptibility for colorectal cancer (CRC) is achievable for a high proportion of familial CRC by exome sequencing. CLINVAR
25645574 ↗ ACG clinical guideline: Genetic testing and management of hereditary gastrointes CLINVAR
26389258 ↗ Cancer Genetics Risk Assessment and Counseling (PDQ®): Health Professional Version. CLINVAR
26467025 ↗ A Standardized DNA Variant Scoring System for Pathogenicity Assessments in Mende CLINVAR
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification crite CLINVAR