Back
NM_002439.5:c.1897-12T>A
p.? · MSH3
ACMG/AMP
0%
complete
Final classification
VUS
MSH3
c.1897-12T>A
p.?
unknown · exon 13i

MSH3 encodes a protein that partners with MSH2 to form MutS beta, a key component of the DNA mismatch repair system that recognizes and corrects errors made during DNA replication. It acts as a tumor suppressor: loss of MSH3 function leads to mismatch repair deficiency, an elevated mutation rate, and increased cancer risk, and the gene is frequently altered in mismatch repair-deficient colorectal cancers. Defects in MSH3 are linked to susceptibility to endometrial cancer, and inherited mutations are associated with an autosomal recessive form of familial adenomatous polyposis, with minimal evidence for a role in Lynch syndrome. Tumors with mismatch repair deficiency, including those involving MSH3, tend to respond well to immune checkpoint inhibitor therapies.

This variant

MSH3 encodes the MSH2-partner protein MutSβ, which supports DNA mismatch repair, and inherited biallelic MSH3 defects are associated with autosomal recessive familial adenomatous polyposis and cancer susceptibility.

Transcript
NM_002439.5
HGVS · transcript:coding
NM_002439.5:c.1897-12T>A
GRCh38
chr5:80767921 T>A
GRCh37
chr5:80063740 T>A
VUS: no adjudicated ACMG/AMP criteria were applied, so no pathogenic, likely pathogenic, benign, or likely benign combination threshold was met.
Classification rationale
VUS
MSH3 c.1897-12T>A unknown · exon 13i

Only PVS1, PM4, and BP3 were adjudicated. No applicable MSH3 VCEP or gene-specific framework was available; the generic ClinGen SVI PVS1 framework was used for the consequence assessment. No applicable MSH3-specific VCEP or gene-specific functional-evidence specification was identified, so generic ACMG/AMP principles were used. No validated variant-specific functional assay with interpretable abnormal or normal results was found; computational SpliceAI output does not satisfy PS3 or BS3. No applicable MSH3-specific VCEP or gene-specific computational-evidence specification was identified, so the supplied generic ACMG/ClinGen-SVI calibration thresholds were used. The intronic consequence selects SpliceAI as the sole PP3/BP4 path; the SpliceAI maximum delta of 0.163 falls between the supplied PP3 supporting and BP4 supporting cutoffs. BP7 is upstream not_applicable because the variant is not synonymous. No applicable MSH3-specific VCEP or gene framework was available, so the supplied generic population thresholds were used. BA1 and BS1 are not met because the observed frequencies remain below 0.05 and 0.01, respectively; PM2 is not met because all-comers and non-cancer frequencies exceed 0.0001. BS2 remains not assessed because homozygote observations lack documented age, health status, inheritance-model context, and penetrance information.

LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_002439.5 · variants mapped to exon structure
MSH3 NM_002439.5
Fetching transcript structure from UCSC…
Applied criteria · 0 applied · 20 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 0

No criteria were applied for this variant.
Assessed · not applied · 8 not met · 12 not assessed
Pathogenic
PVS1 Not met: c.1897-12T>A is a noncanonical intronic SNV, and SpliceAI max delta 0.163 does not establish a loss-of-function splice consequence.
PS2 Not assessed: no documented affected proband with confirmed de novo status or parental genotyping is available.
PS3 Not assessed: no validated variant-specific functional assay with interpretable abnormal results is available for NM_002439.5:c.1897-12T>A.
PS4 Not assessed: no case-control enrichment, odds ratio, likelihood ratio, or affected-case excess is reported for the exact variant.
PM2 Not met: gnomAD v4.1 overall AF is 0.00033546, exceeding the generic PM2 threshold of 0.0001.
PM3 Not assessed: no affected proband, second pathogenic MSH3 variant, or confirmed trans phase is documented for NM_002439.5:c.1897-12T>A.
PM6 Not assessed: no affected-proband de novo observation is reported without parental testing.
PP1 Not assessed: no informative meioses or affected-relative segregation data are reported for the variant.
PP3 Not met: SpliceAI maximum delta 0.163 is below the >=0.2 PP3 supporting threshold.
PP4 Not assessed: no patient phenotype or disease-specific clinical presentation is documented for this exact variant.
PP5 Not met: ClinVar lists 0 expert-panel submissions for the exact variant, so ordinary laboratory assertions cannot support PP5.
Benign
BA1 Not met: the highest observed allele frequency is 0.0061081, below the generic BA1 threshold of 0.05.
BS1 Not met: the maximum supported subgroup frequency is 0.0061081, below the generic BS1 threshold of 0.01.
BS2 Not assessed: three gnomAD v4.1 homozygotes are reported, but adult health, inheritance model, and penetrance are undocumented.
BS3 Not assessed: no validated variant-specific functional assay with interpretable normal results is available for NM_002439.5:c.1897-12T>A.
BS4 Not assessed: no phenotype-qualified unaffected relatives with negative variant testing are documented.
BP2 Not assessed: no relevant individual with a second pathogenic variant and documented cis or trans phase is reported for this variant.
BP4 Not met: SpliceAI maximum delta 0.163 exceeds the <=0.1 BP4 supporting threshold.
BP5 Not assessed: no affected individual has documented alternative molecular etiology explaining the phenotype alongside this variant.
BP6 Not met: ClinVar lists 0 expert-panel submissions, and ordinary laboratory Benign or Likely benign assertions cannot support BP6.
N/A · 8 PS1 · PM1 · PM4 · PM5 · PP2 · BP1 · BP3 · BP7
Research & evidence
Population frequency
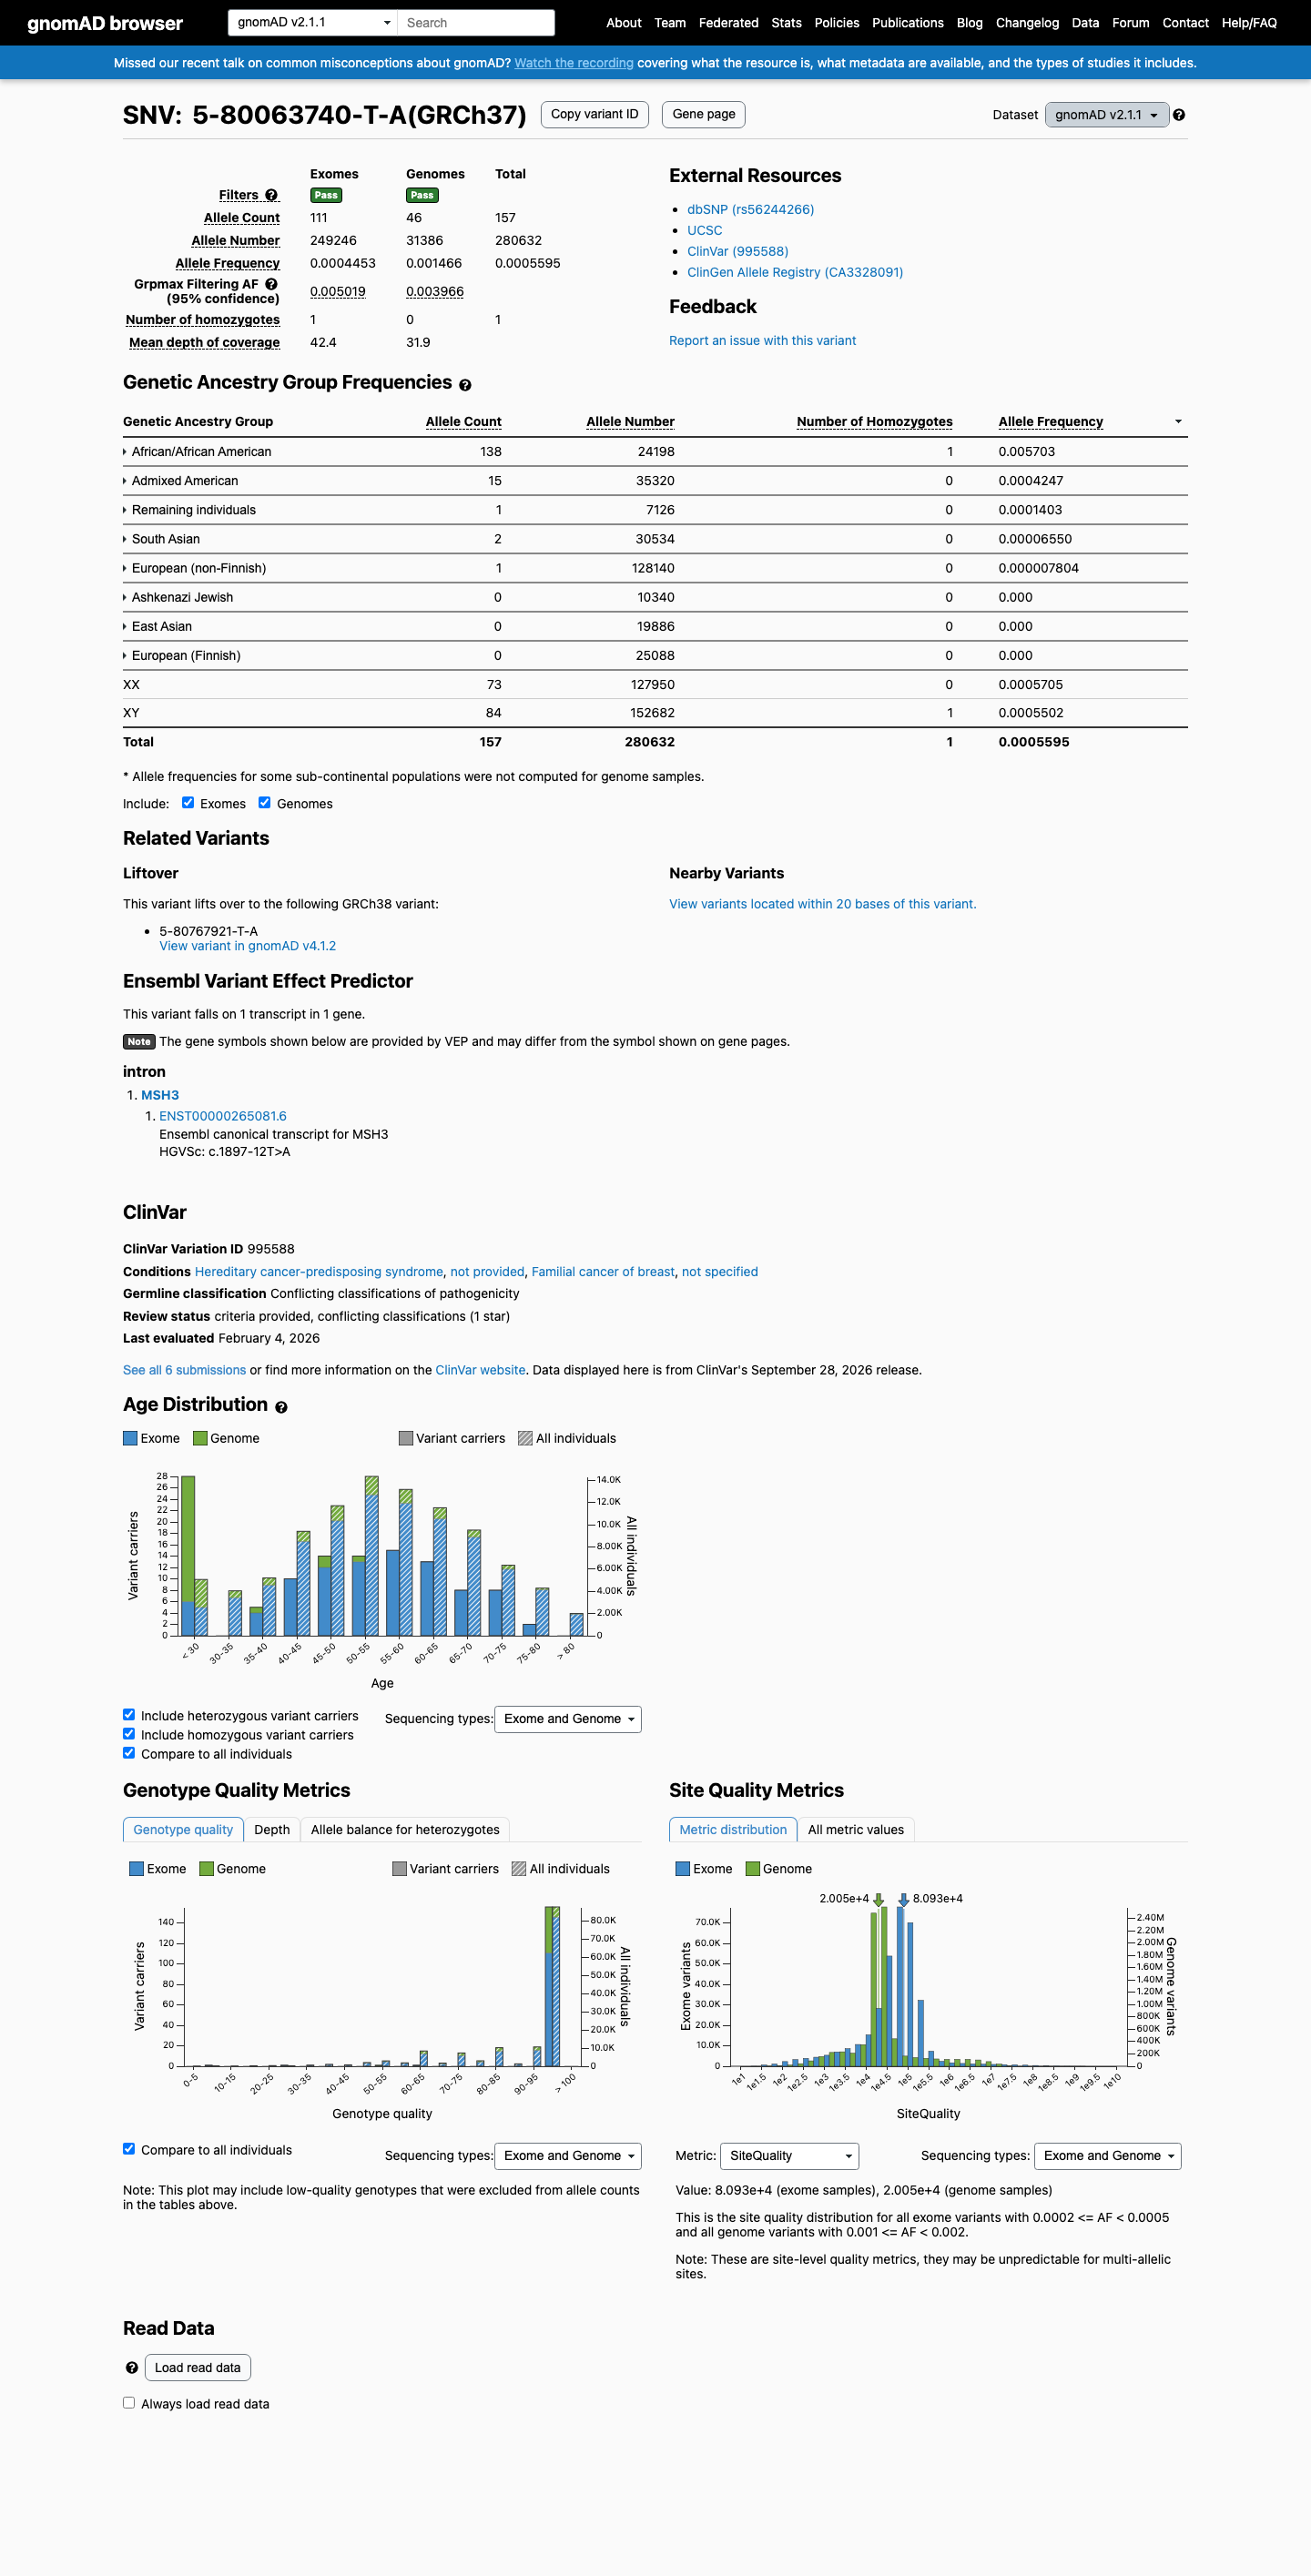
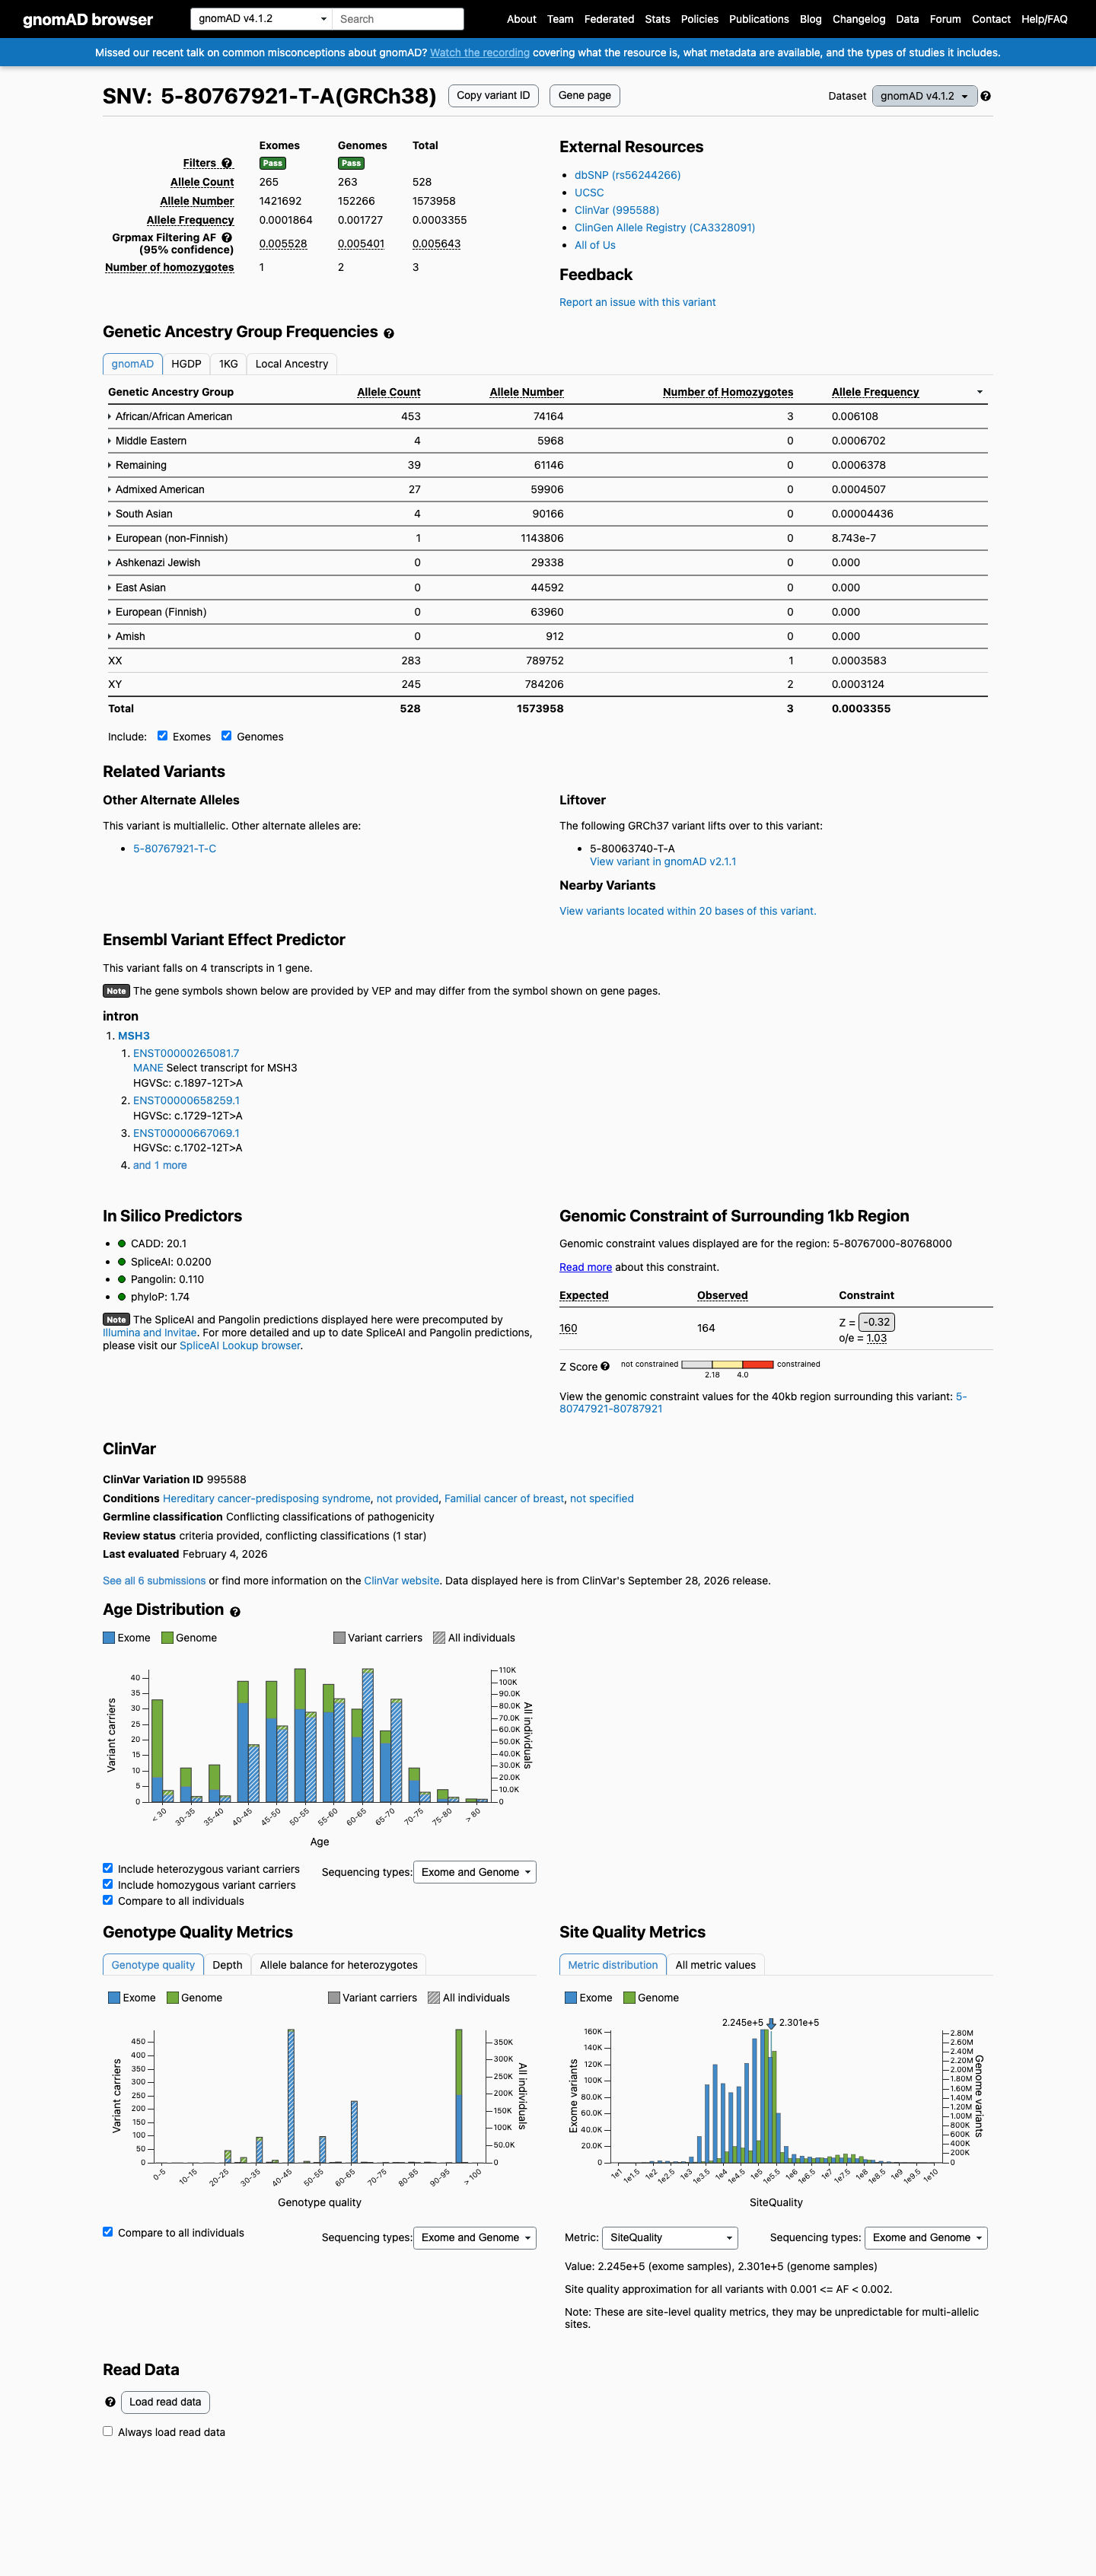
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 0.00033546; MAF= 0.03355%, 528/1573958 alleles, homozygotes = 3) and has highest observed frequency in the African/African American population (AF= 0.00610808; MAF= 0.61081%, 453/74164 alleles, homozygotes = 3); grpmax FAF= 0.00564283.
v2.1
This variant is present in gnomAD v2.1 (AF= 0.000559452; MAF= 0.05595%, 157/280632 alleles, homozygotes = 1) and has highest observed frequency in the African/African American population (AF= 0.00570295; MAF= 0.57030%, 138/24198 alleles, homozygotes = 1); grpmax FAF= 0.00501897.
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.034% · 528 / 1,573,958
3 hom · FAF 0.56%
African/African American
453 / 74,164
0.61%
3 hom
Middle Eastern
4 / 5,968
0.067%
Remaining individuals
39 / 61,146
0.064%
Admixed American
27 / 59,906
0.045%
South Asian
4 / 90,166
0.0044%
European (non-Finnish)
1 / 1,143,806
8.7e-05%
+ 4 not observed (European (Finnish), Amish, East Asian, Ashkenazi Jewish)
gnomAD v2.1
0.056% · 157 / 280,632
1 hom · FAF 0.5%
African/African American
138 / 24,198
0.57%
1 hom
Admixed American
15 / 35,320
0.042%
Remaining individuals
1 / 7,126
0.014%
South Asian
2 / 30,534
0.0066%
European (non-Finnish)
1 / 128,140
0.00078%
+ 3 not observed (Ashkenazi Jewish, East Asian, European (Finnish))
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
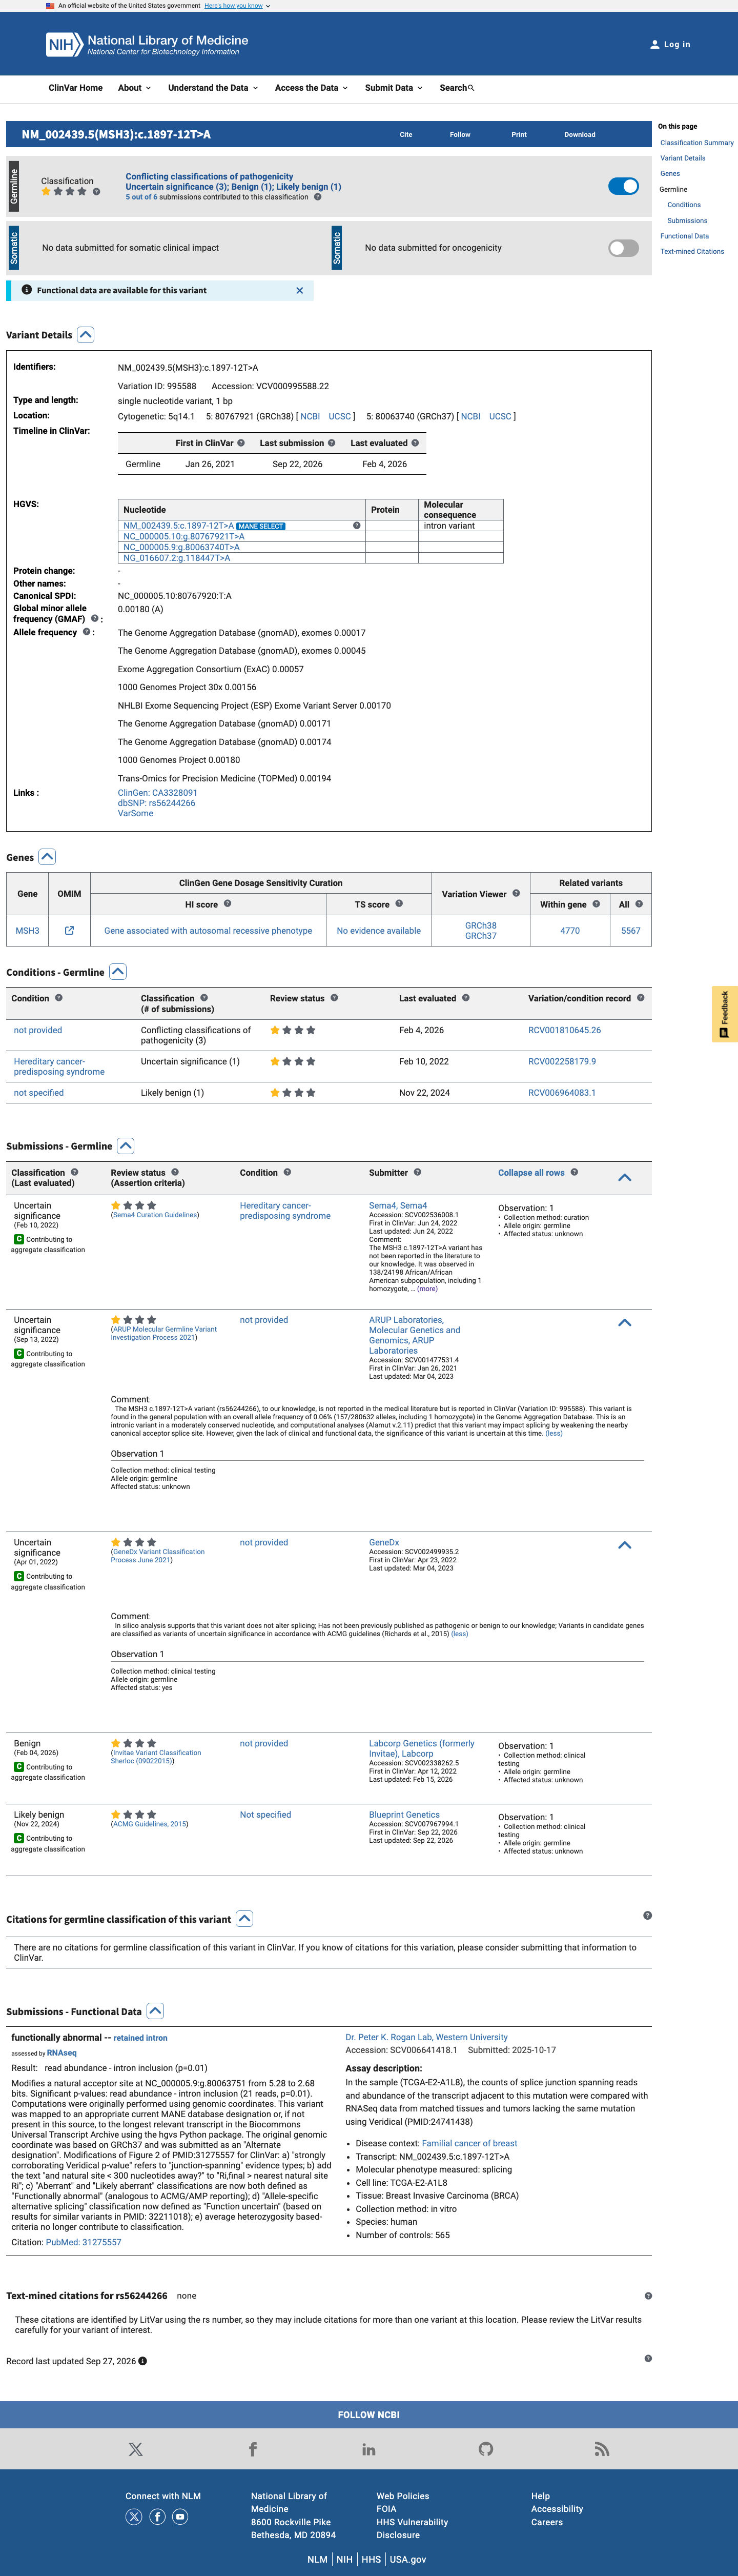
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Uncertain significance (2 clinical laboratories) and as Benign (1 clinical laboratory) and as Likely benign (1 clinical laboratory). (ClinVarID = 995588)
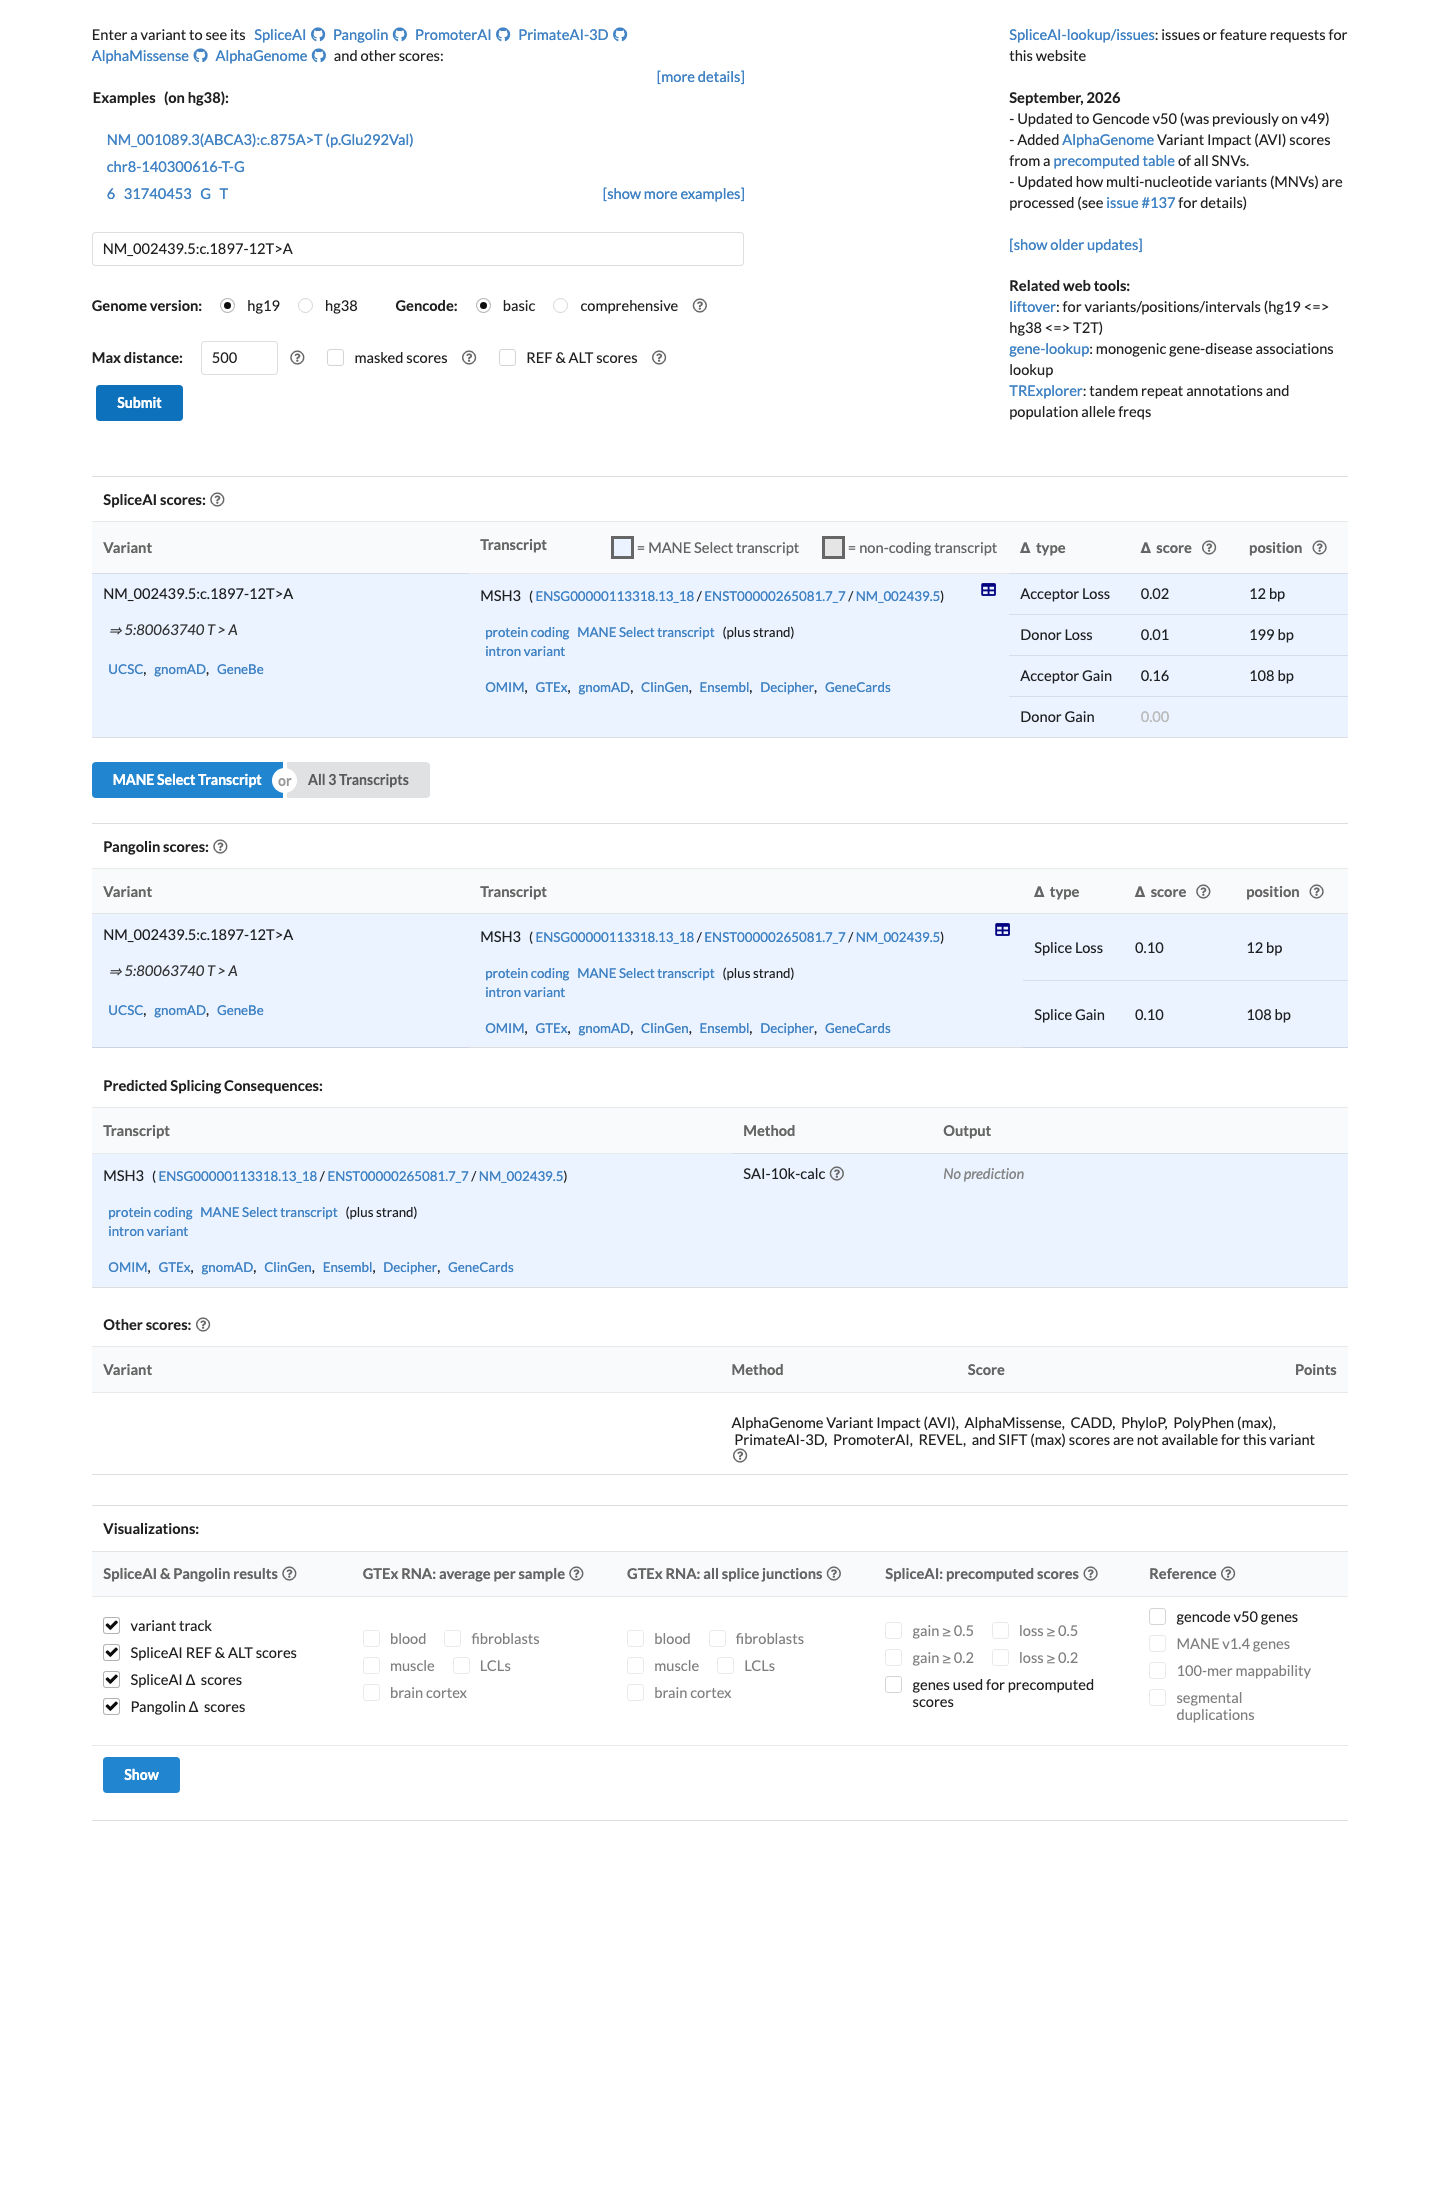
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.16).
Functional No data
No calibrated functional assay or RNA evidence was identified for this variant.
OncoKB ↗
COSMIC screenshot
COSMIC
Somatic evidence
COSMIC
This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant cancer hotspot.
COSMIC ↗
Sources & reference links
7Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Triaged references · 4 PMIDs not cited in assessment
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology. CLINVAR
31275557 ↗ Pan-cancer repository of validated natural and cryptic mRNA splicing mutations. CLINVAR
25394175 ↗ A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment. CLINVAR
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification criteria. CLINVAR