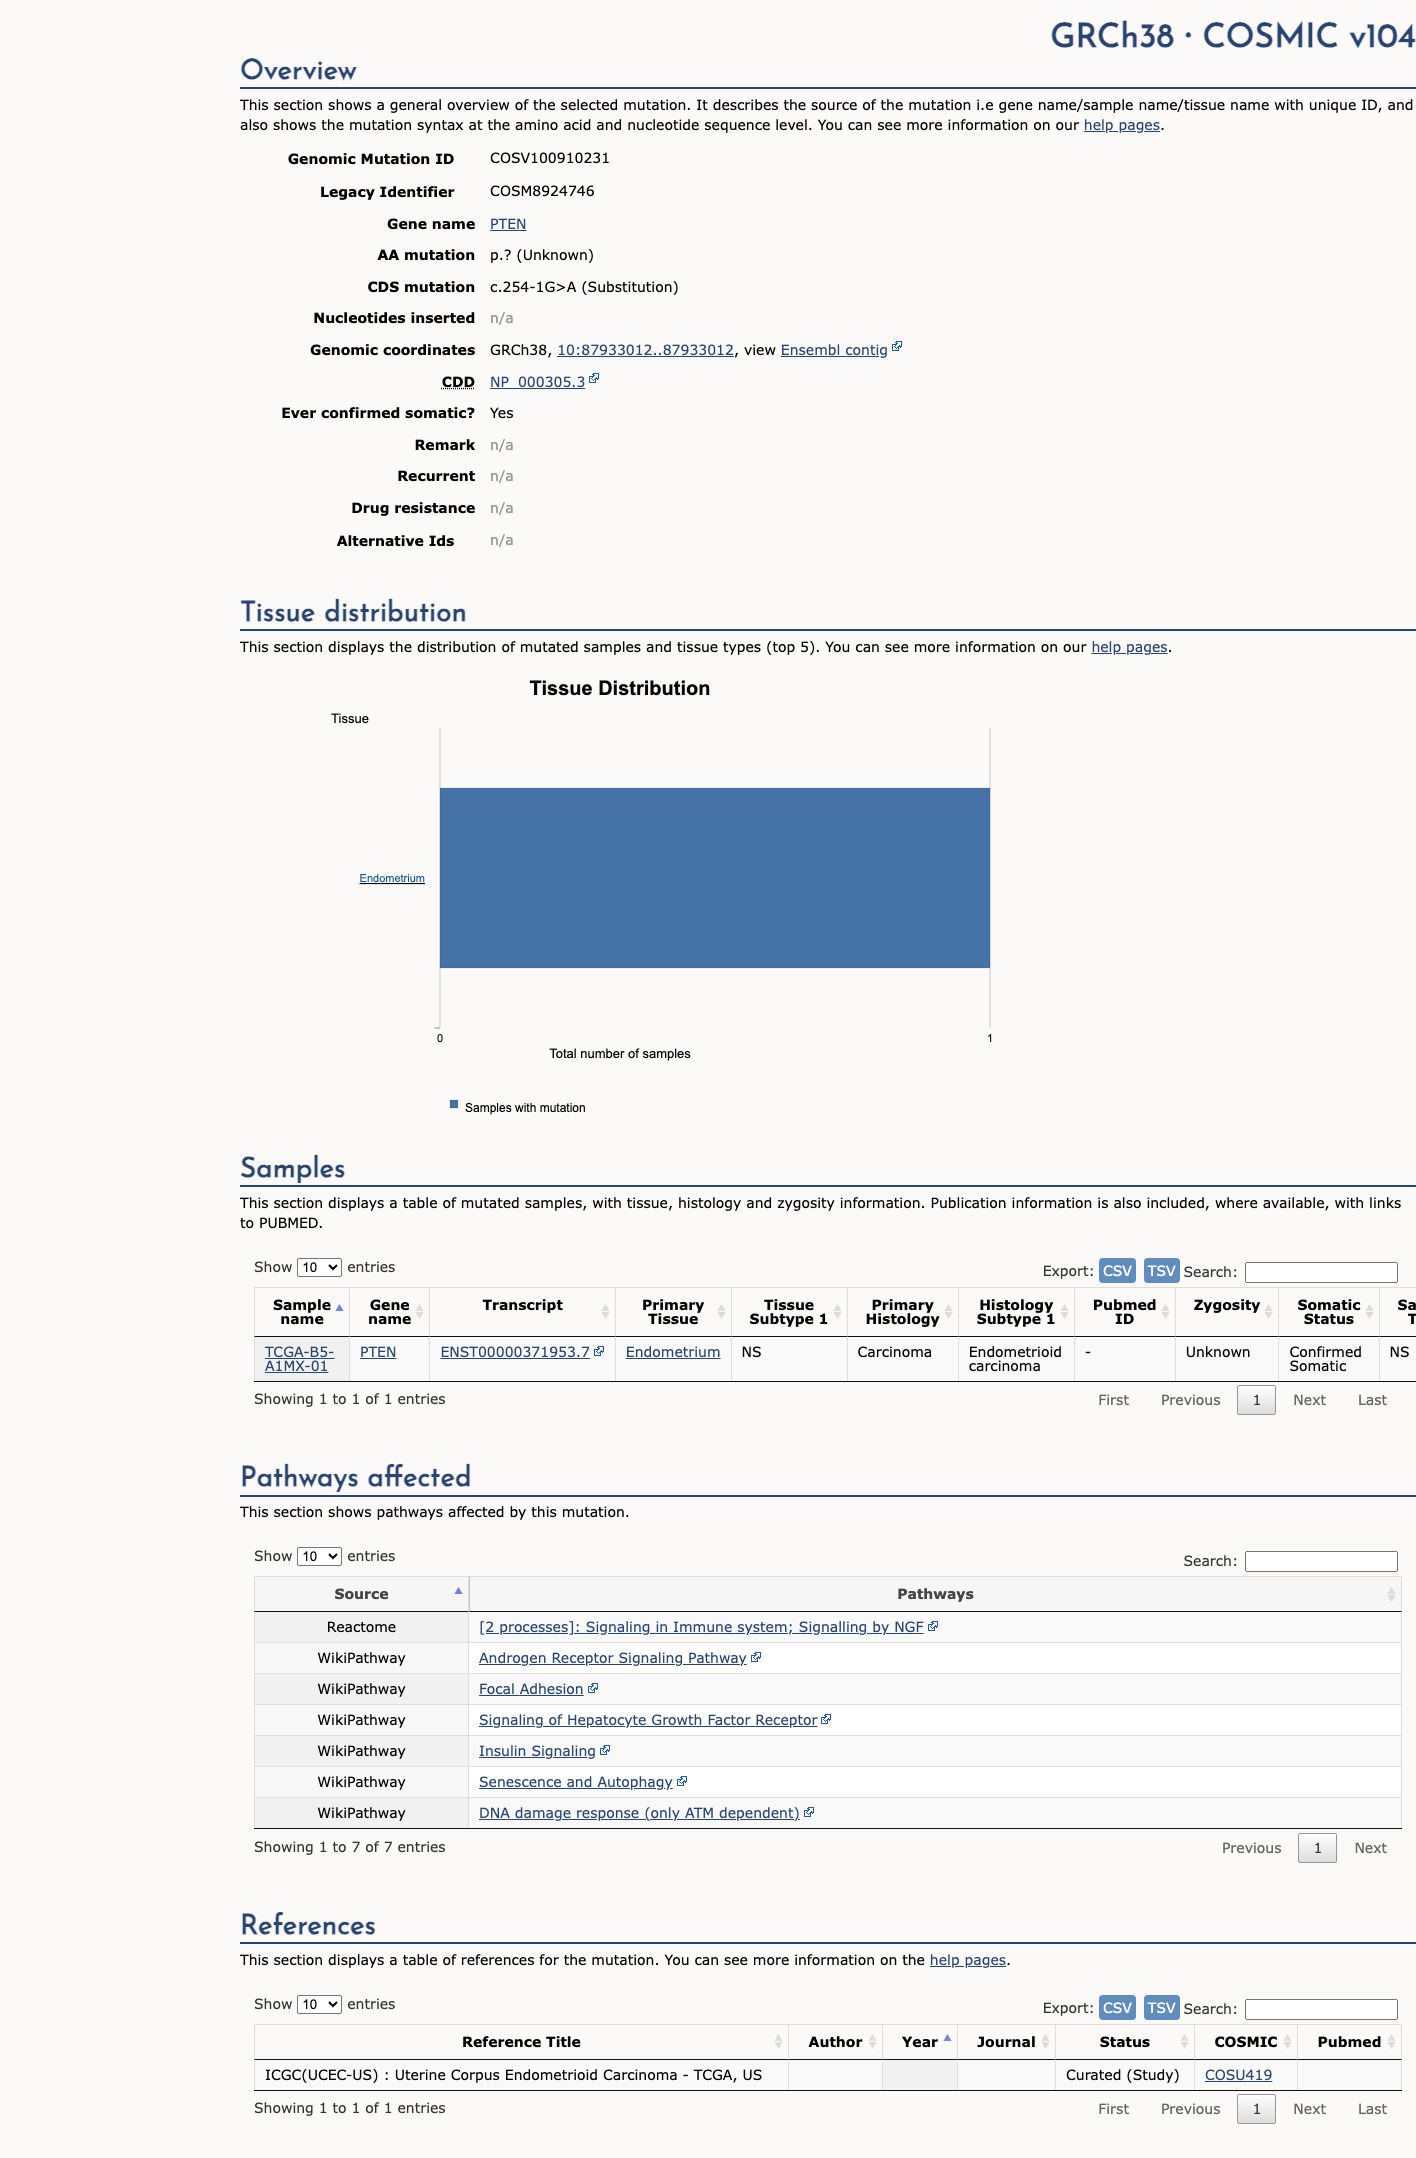
PTEN is a tumor suppressor gene that encodes a phosphatase converting the lipid messenger PIP3 back to PIP2 at the cell membrane, thereby restraining the AKT/mTOR signaling pathway that drives cell growth, proliferation, and survival. It is one of the most frequently mutated genes across many types of human cancer, and its loss promotes unchecked cell growth, survival, and genomic instability, partly through impaired DNA repair. Germline loss-of-function variants in PTEN cause Cowden syndrome, an inherited cancer predisposition disorder associated with elevated risk of breast and thyroid cancer.
This variant
This PTEN splice-site variant is relevant to PTEN-related cancer predisposition because loss of PTEN tumor-suppressor function increases AKT/mTOR signaling and promotes unchecked cell growth.
Transcript
NM_000314.8
HGVS · transcript:coding
NM_000314.8:c.254-1G>A
GRCh38
chr10:87933012 G>A
GRCh37
chr10:89692769 G>A
Likely Pathogenic: PVS1 (very strong) plus PM2 (supporting) satisfy Rule20 of the PTEN Expert Panel framework.
Classification rationale
PVS1PM2Likely Pathogenic
PTEN c.254-1G>Acanonical_splice · exon 4i
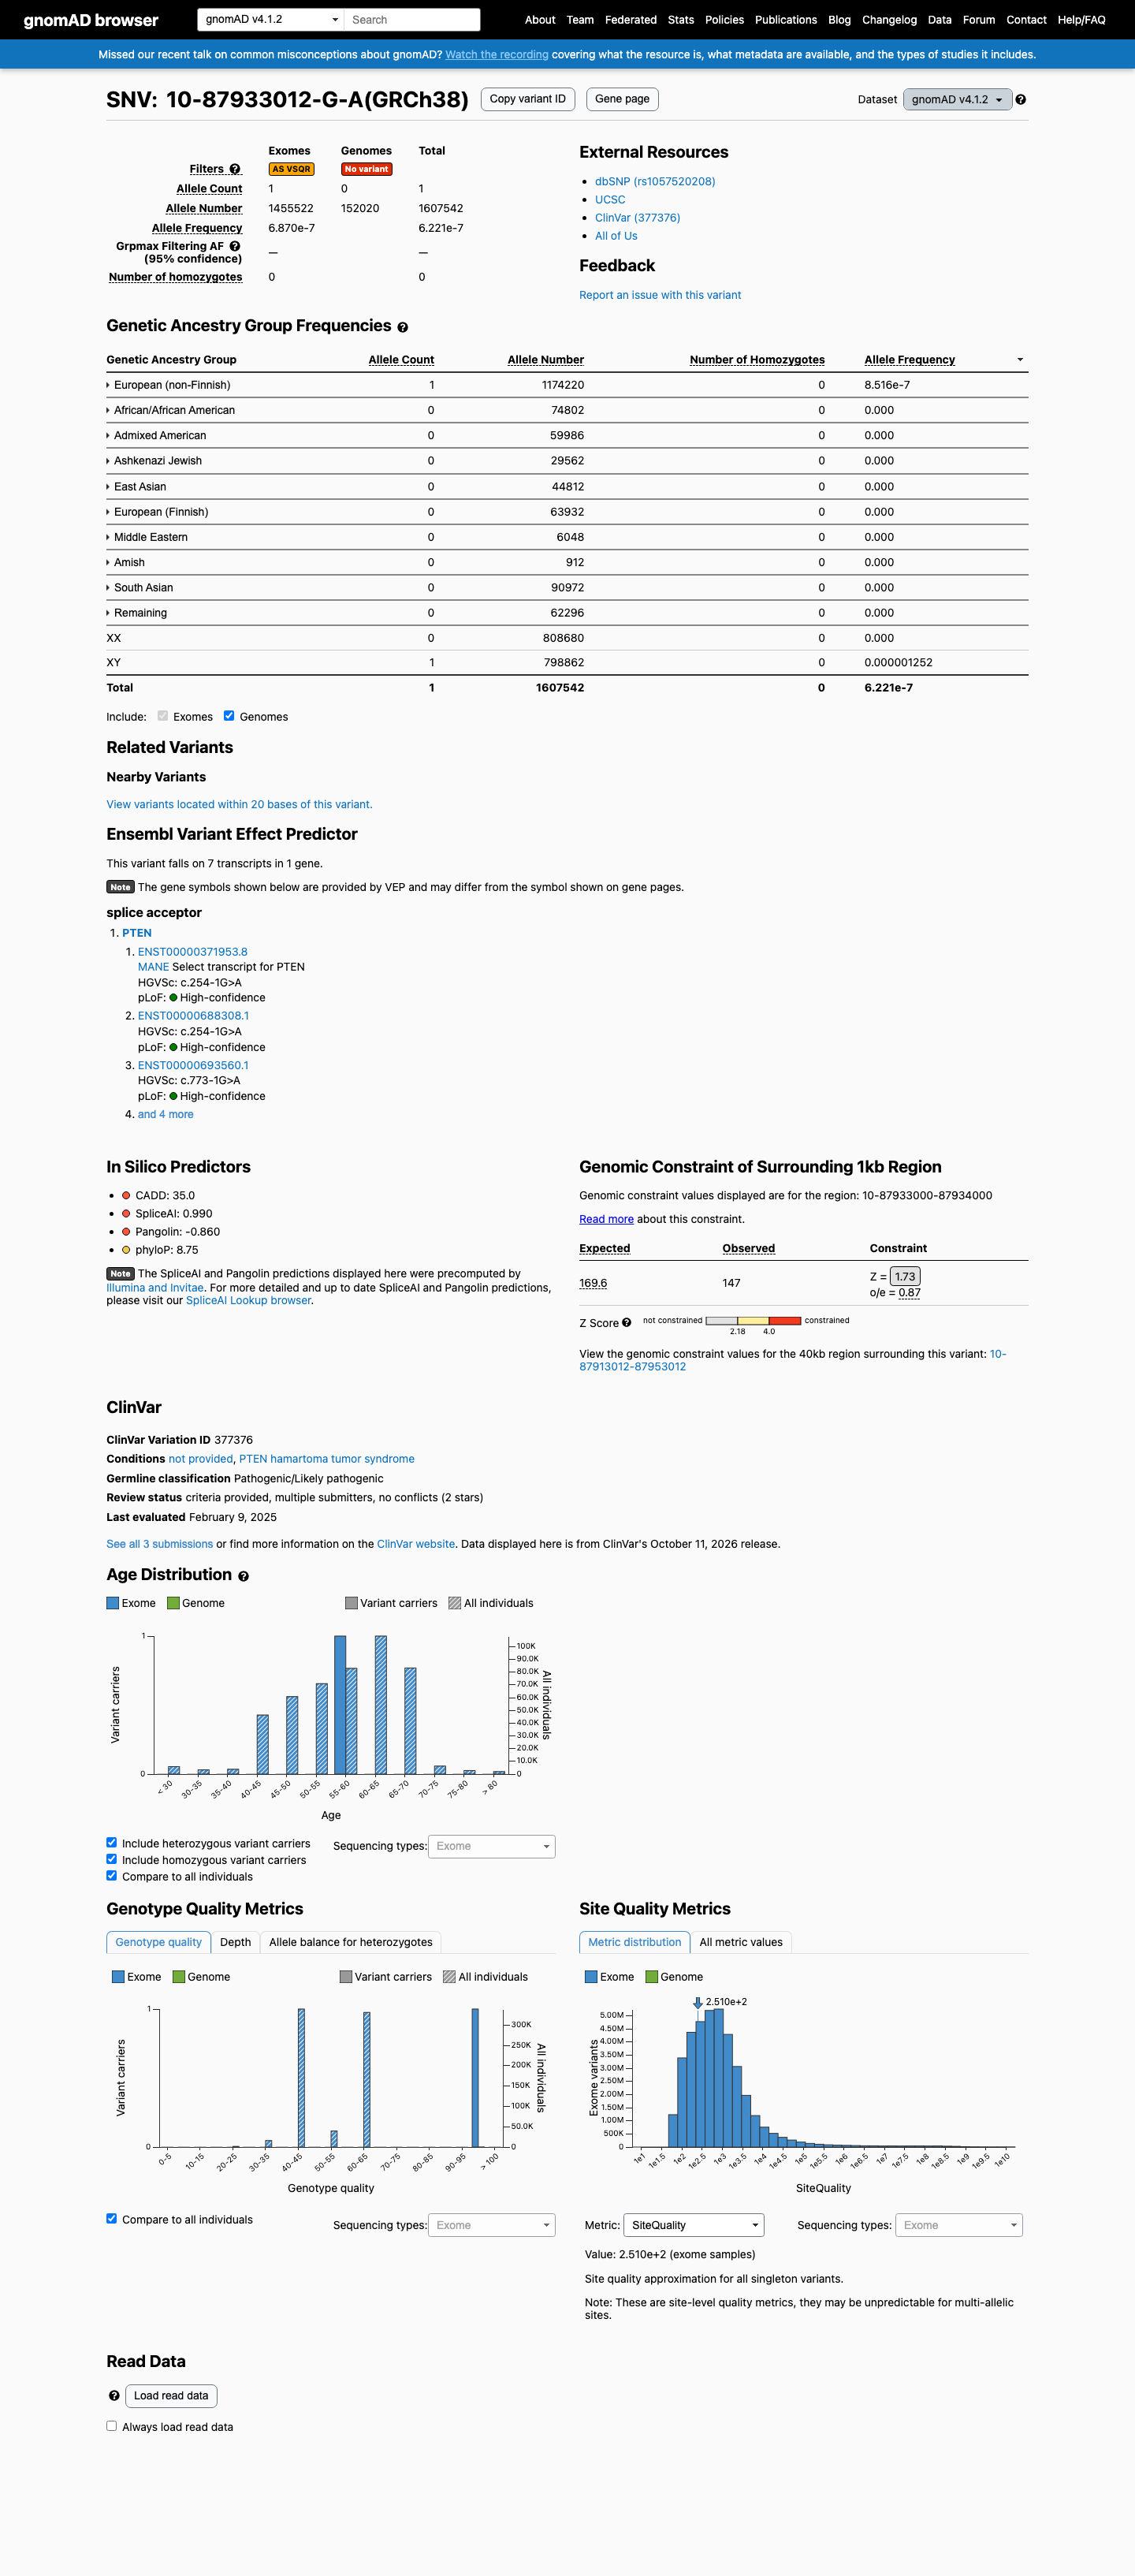
PVS1 very strong: the canonical -1 splice-site change is predicted to cause a frameshifted exon-5 skipping transcript upstream of the NMD threshold. PM2 supporting: the gnomAD v4.1 allele frequency is below the PTEN VCEP threshold, with zero homozygotes reported.
PVS1 + PM2→Likely Pathogenic
LYFE Sciences is an AI system, and it can make mistakes. Criteria
may be applied incorrectly, sources may be misread, and a confident-looking
classification can still be wrong. Double-check every criterion and
its underlying evidence before relying on any call.
Gene diagram
· NM_000314.8 · variants mapped to exon structure
PTENNM_000314.8
Fetching transcript structure from UCSC…
Exons
—
Transcript span
—
Strand
—
Variants mapped
—
Source
UCSC ncbiRefSeqCurated
All variants in PTEN—click a row to locate it on the plot · use the link column to open its page
Protein
Location
Classification
Link
Applied criteria · 2 applied · 15 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
✓
PVS1very strongPathogenic
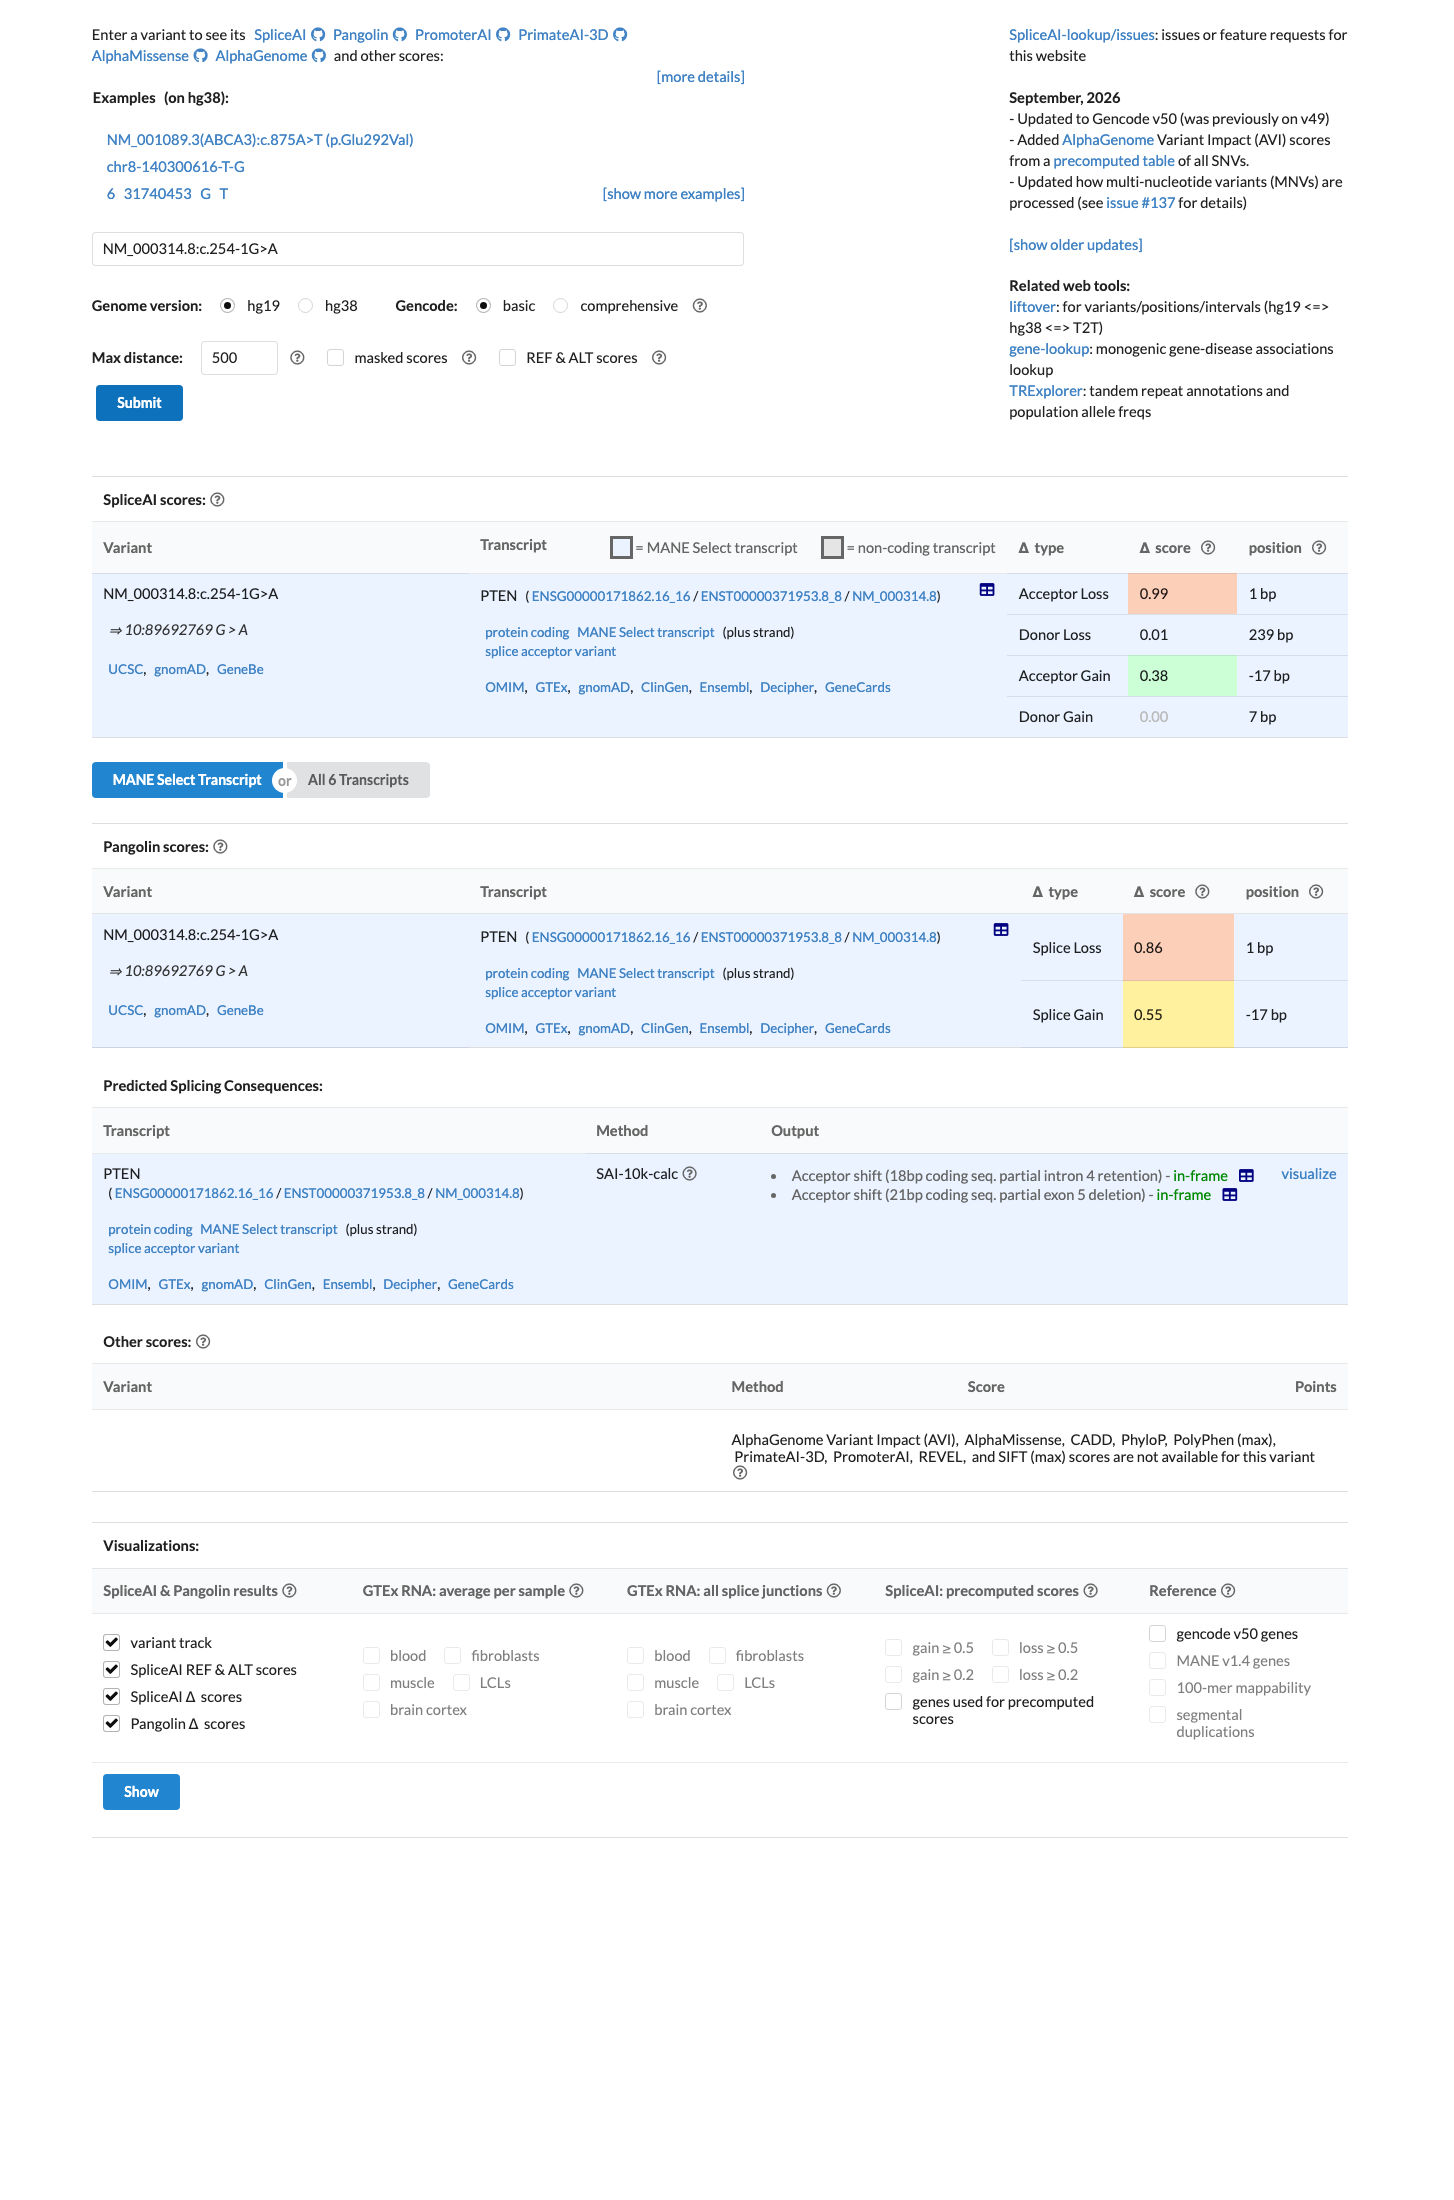
Met at very strong: canonical -1 acceptor disruption causes a predicted exon-5 frameshift after loss of 239 coding nucleotides and lies 5' of the PTEN c.1121 NMD threshold.
pvs1_variant_assessment records NM_000314.8:c.254-1G>A as a canonical splice variant at the acceptor (-1), affecting exon 5 of 9 in the biologically relevant transcript.pvs1_variant_assessment records exon 5 as c.254_492, with 239 coding nucleotides; exon skipping removes 239 nt (239 % 3 = 2) and shifts the reading frame.PVS1_DecisionTree_PTEN.pdf, page 1: the GT-AG +/-1,2 splice-site branch assigns PVS1 when exon skipping or cryptic splice-site use disrupts the reading frame and is predicted to undergo NMD, the disruption is at or 5' to p.D375 (c.1121), and the exon is present in NM_000314.8.
Met at supporting strength: gnomAD v4.1 total AF is 6.220677282459805e-07, below the <0.00001 PM2 threshold, with 0 homozygotes.
ClinGen PTEN VCEP Version 3.2 PM2 rule: databases present at <0.00001 (0.001%) allele frequency in gnomAD or another large sequenced population; if multiple alleles are present within any subpopulation, that subpopulation allele frequency must be <0.00002 (0.002%).gnomAD v4.1 reports total AF 6.220677282459805e-07 (1/1607542 alleles), highest listed subpopulation AF 8.516291665956976e-07 (1/1174220 alleles), and 0 homozygotes.gnomAD v2.1 reports the variant absent; the available gnomAD v2.1 non-cancer and v3.1 non-cancer queries also report absence.
Assessed · not applied
· 1 not met · 14 not assessed
Pathogenic
PS1Not assessed: SpliceAI is 0.993, but no known pathogenic comparator at the same nucleotide position is available for the VCEP's required equal-or-greater comparison.
PS2Not assessed: no documented proband-level de novo observation, parental testing, or family-history status is available to evaluate the PS2 thresholds.
PS3Not assessed: no qualifying splice assay result is available, and the VCEP's phosphatase threshold applies to missense assay data absent for this canonical splice-site variant.
PS4Not assessed: no variant-specific proband phenotype, specificity score, or case-control enrichment statistic is available for applying the PTEN VCEP PS4 rule.
PM6Not assessed: no presumed de novo proband observation, family-history status, or parental-confirmation details are documented for applying PM6.
PP1Not assessed: no affected relatives or informative meioses are documented, so the PP1 segregation thresholds cannot be applied.
PP3Not assessed: the rule requires concordant SpliceAI and VarSeak predictions, but only SpliceAI is available, with maximum delta 0.993.
Benign
BA1Not assessed: gnomAD filtering allele frequency is unavailable, although total AF is 6.220677282459805e-07 versus the >0.00056 BA1 threshold.
BS1Not assessed: gnomAD filtering allele frequency is unavailable, while total AF is 6.220677282459805e-07 below the 0.0000043 BS1-supporting lower bound.
BS2Not met: gnomAD v4.1 reports 0 homozygotes, but the rule requires at least one homozygous observation in a healthy or PHTS-unaffected individual.
BS3Not assessed: no benign splicing assay demonstrates no impact, and the VCEP's phosphatase criterion applies to missense assay data absent for this canonical splice-site variant.
BS4Not assessed: no affected-family non-segregation observation is documented for evaluating BS4 at supporting or strong strength.
BP2Not assessed: no documented pathogenic PTEN variant in cis or trans, or phase result, establishes either VCEP BP2 pathway.
BP4Not assessed: the rule requires concordant benign SpliceAI and VarSeak predictions, but SpliceAI's maximum delta is 0.993 and VarSeak is unavailable.
BP5Not assessed: no two cases with an alternate highly penetrant molecular diagnosis and documented non-overlapping PTEN history meet the BP5 requirements.
This variant is present in gnomAD v4.1 (AF= 6.22068e-07; MAF= 0.00006%, 1/1607542 alleles, homozygotes = 0) and has highest observed frequency in the European (non-Finnish) population (AF= 8.51629e-07; MAF= 0.00009%, 1/1174220 alleles, homozygotes = 0).
v2.1
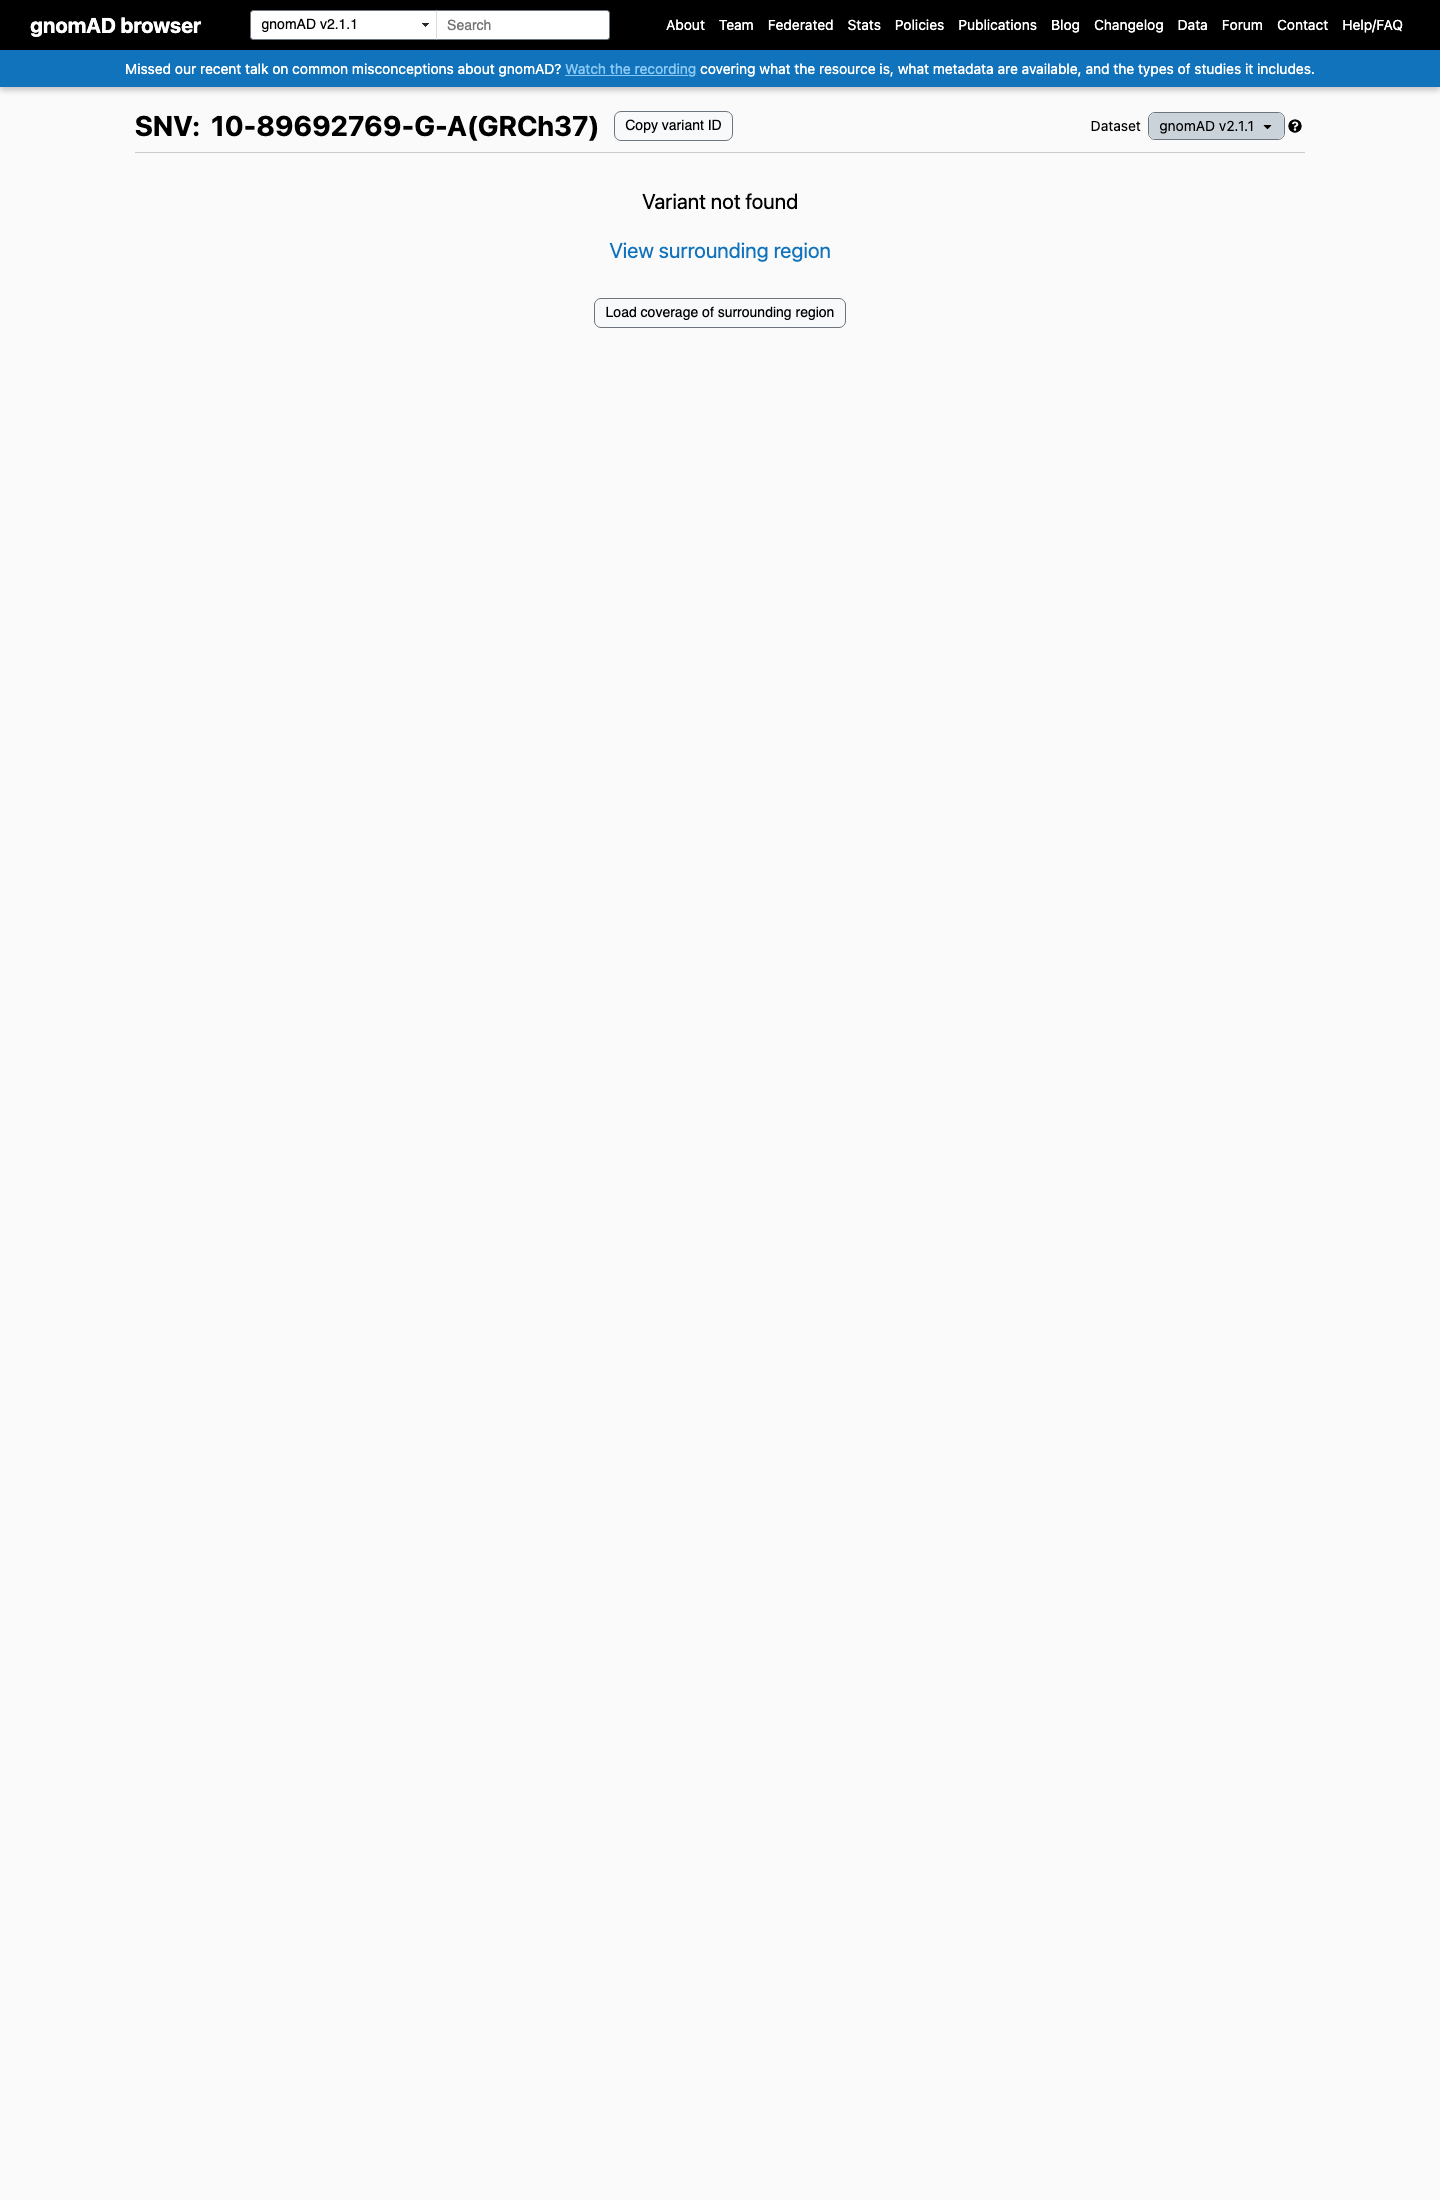
Absent from gnomAD v2.1.
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
6.2e-05%
· 1 / 1,607,542
0 hom
European (non-Finnish)
1 / 1,174,220
8.5e-05%
+ 9 not observed (Remaining individuals, Admixed American, European (Finnish), Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American)
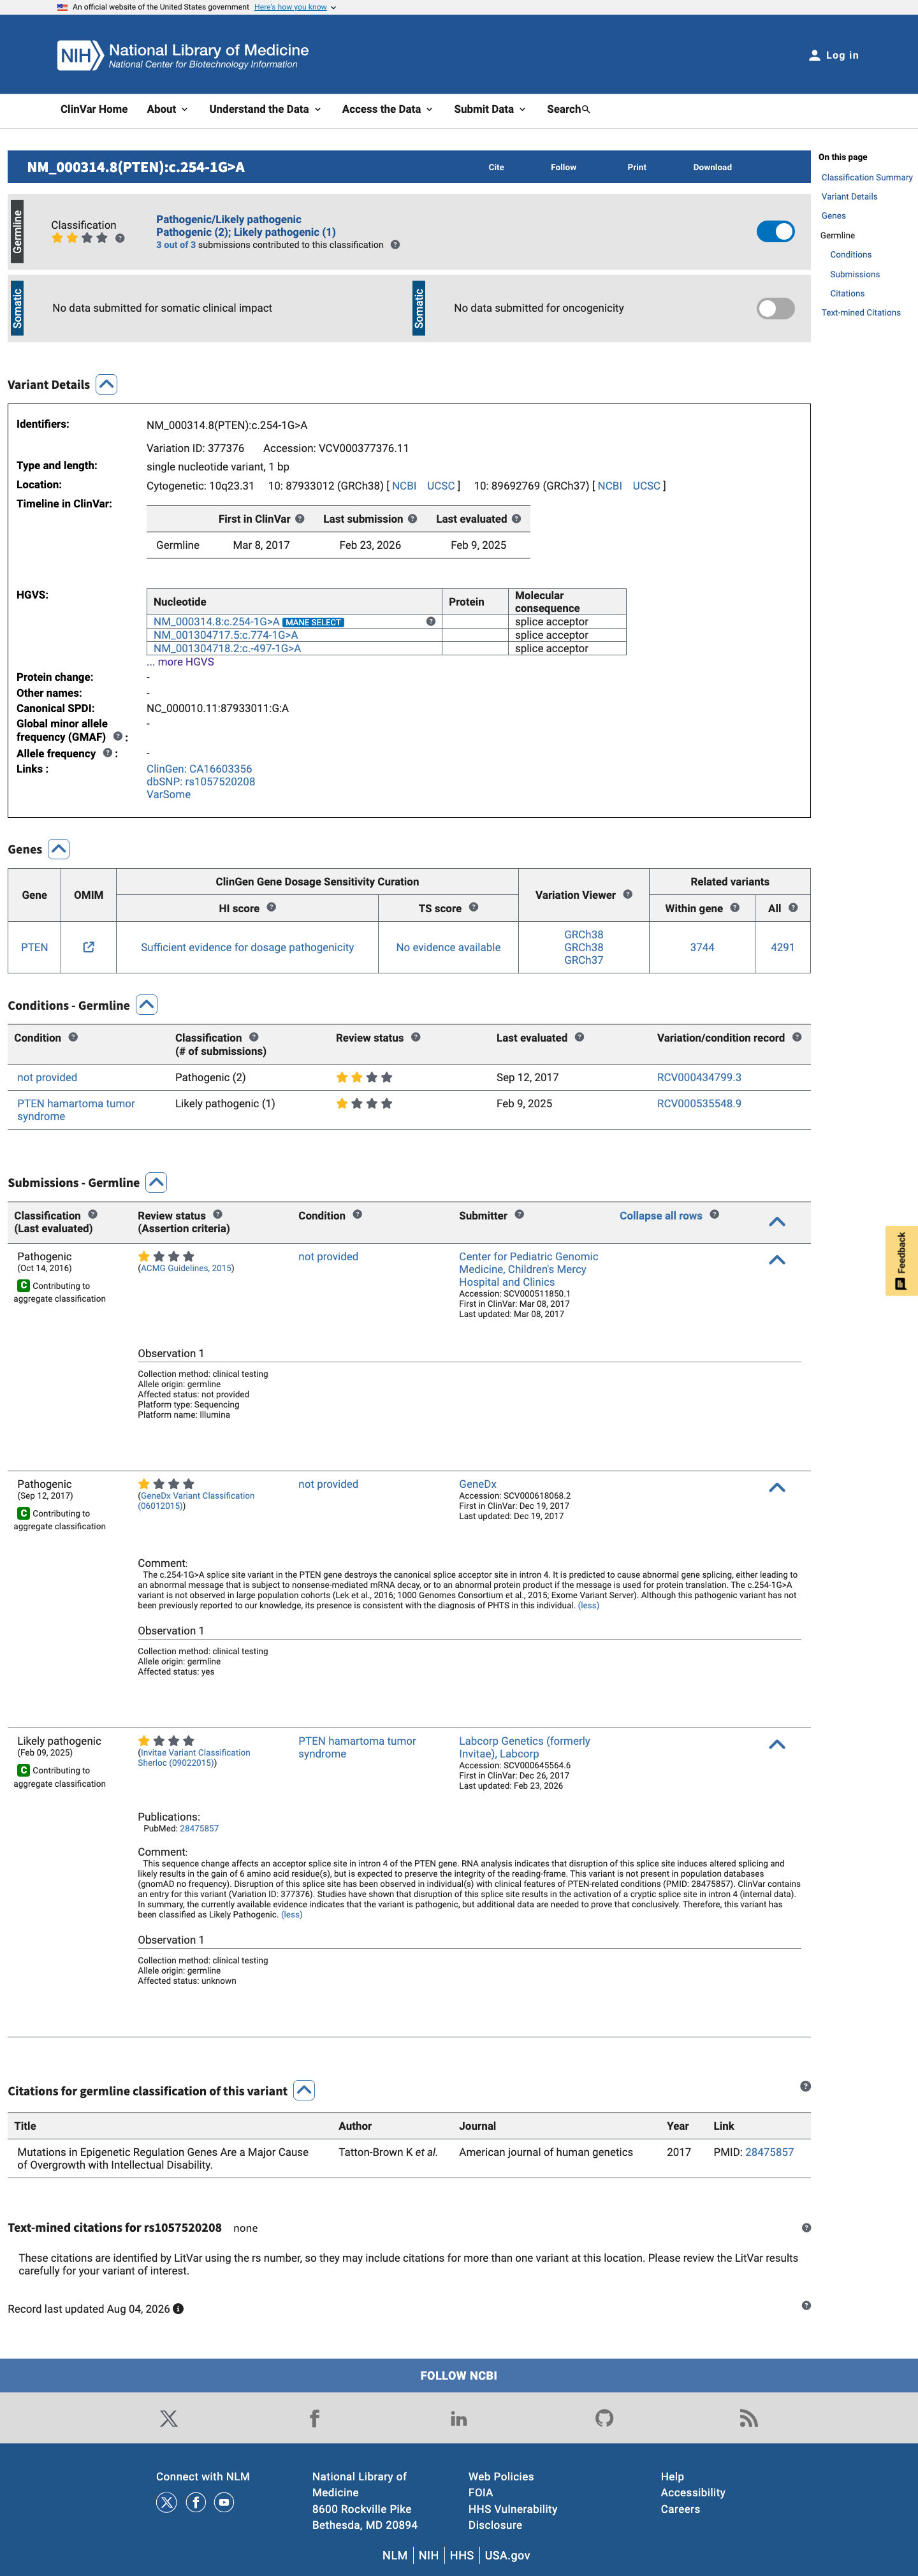
This variant has been reported in ClinVar as Pathogenic (2 clinical laboratories) and as Likely pathogenic (1 clinical laboratory). (ClinVarID = 377376)
Triaged references · 7 PMIDs not cited in assessment
25741868 ↗Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.CLINVAR
28475857 ↗Mutations in Epigenetic Regulation Genes Are a Major Cause of Overgrowth with Intellectual Disability.CLINVAR
17392385 ↗American Cancer Society guidelines for breast screening with MRI as an adjunct to mammography.CLINVAR
23519317 ↗Clinical genetics evaluation in identifying the etiology of autism spectrum disorders: 2013 guideline revisions.CLINVAR
23788249 ↗ACMG recommendations for reporting of incidental findings in clinical exome and genome sequencing.CLINVAR
25190698 ↗Genetic/familial high-risk assessment: breast and ovarian, version 1.2014.CLINVAR
27854360 ↗Recommendations for reporting of secondary findings in clinical exome and genome sequencing, 2016 update (ACMG SF v2.0): a policy statement of the American College of Medical Genetics and Genomics.CLINVAR