Back
NM_006231.4:c.6380G>A
p.Arg2127Gln · POLE
León-Castillo et al. 2020 custom POLE framework · vleon-castillo-2020-custom-framework-v1
0%
complete
Final classification
VUS
POLE
c.6380G>A
p.Arg2127Gln
missense · exon 46

POLE encodes the catalytic subunit of DNA polymerase epsilon, the enzyme that replicates the leading strand of DNA during cell division and participates in DNA repair. It contains a proofreading domain that corrects replication errors, keeping the accumulation of mutations in check. Germline mutations in POLE cause polyposis and predispose to colorectal cancer, and are also linked to a rare syndrome of facial dysmorphism, immunodeficiency, livedo, and short stature. Somatic mutations, particularly in the proofreading domain, occur in colorectal and endometrial cancers, where they drive an ultra-mutated tumor phenotype and are associated with better responses to immune checkpoint inhibitors.

This variant

This POLE missense variant affects a gene whose proofreading function maintains replication fidelity and whose germline alterations are associated with polyposis and colorectal-cancer predisposition.

Transcript
NM_006231.4
HGVS · transcript:coding
NM_006231.4:c.6380G>A
GRCh38
chr12:132626268 C>T
GRCh37
chr12:133202854 C>T
VUS: no adjudicated pathogenic or benign criterion is met, so the local POLE framework satisfies no final classification combination.
Classification rationale
VUS
POLE c.6380G>A missense · exon 46

None of PS2, PM6, PP1, or BS4 can be assessed because the available materials contain no case-level proband, parental, pedigree, affected-relative, meiosis, or segregation data. PS4 is not met under the local POLE framework because the exact variant is absent from the governing recurrence sources and is not in the framework's eligible pathogenic set. PP4 and BP5 are not assessed because the supplied case evidence lacks the required phenotype/family-history or alternate-molecular-basis information. PP5 and BP6 are not met because the exact ClinVar record contains only Uncertain significance clinical-laboratory submissions and no expert-panel classification. Population evidence does not support BA1, BS1, or PM2 under the supplied generic thresholds. BS2 remains not assessed because aggregate population counts do not document a healthy adult carrier or establish the required inheritance, penetrance, and age-of-onset context.

LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_006231.4 · variants mapped to exon structure
POLE NM_006231.4
Fetching transcript structure from UCSC…
Applied criteria · 0 applied · 24 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 0

No criteria were applied for this variant.
Assessed · not applied · 10 not met · 14 not assessed
Pathogenic
PS1 Not met: no previously established pathogenic variant producing the same p.Arg2127Gln amino-acid change was identified.
PS2 Not assessed: no proband-level parental testing, disease phenotype, family-history, or confirmed maternity and paternity are documented.
PS3 Not assessed: no variant-specific validated functional assay or reproducible clinical-laboratory evidence was identified for p.Arg2127Gln.
PS4 Not met: p.Arg2127Gln is absent from the governing recurrence table and lacks the required COSMIC-plus-TCGA count >=10 and established hotspot designation.
PM1 Not met: p.Arg2127Gln is absent from the framework's complete exact-variant PM1 lists, including all five hotspots and recurrent exonuclease-domain substitutions.
PM2 Not met: the variant is present in controls, with 1KG:CHB AF 0.00490196 exceeding the PM2 threshold of <=0.0001.
PM3 Not assessed: no recessive disease context, affected-proband observation, pathogenic allele, or parental/offspring phase testing is documented.
PM5 Not assessed: no defensible pathogenic alternate missense substitution at POLE Arg2127 was available for comparison.
PM6 Not assessed: no affected proband, family-history, parental-testing, or unconfirmed-parentage evidence supports an assumed de novo occurrence.
PP1 Not assessed: no affected relatives, informative meioses, variant observations, or phenotype-consistent co-segregation data are documented.
PP2 Not assessed: no validated gene-level evidence establishes both a low benign missense rate and a common missense disease mechanism for POLE.
PP3 Not met: REVEL was 0.313, within the generic missense gray zone of 0.250–0.750 and below the PP3 threshold of >0.750.
PP4 Not assessed: no patient phenotype or family history sufficiently specific for a single-etiology disease was available for comparison.
PP5 Not met: ClinVar reports p.Arg2127Gln as Uncertain significance, with zero expert-panel submissions and no exact pathogenic classification.
Benign
BA1 Not met: the highest reported ancestry-specific AF is 0.00490196, below the BA1 stand-alone threshold of 0.05.
BS1 Not met: the maximum reported AF is 0.00490196, below the generic BS1 threshold of 0.01.
BS2 Not assessed: zero homozygotes are reported, but no documented healthy adult carrier or penetrance and age-of-onset assessment is available.
BS3 Not assessed: no variant-specific validated functional assay demonstrated preserved protein function or normal splicing for p.Arg2127Gln.
BS4 Not assessed: no affected relatives with documented absence of the variant, phenotype review, or informative non-segregation data are available.
BP1 Not assessed: no validated gene-level evidence shows that POLE disease is caused primarily by truncating variants.
BP2 Not assessed: no qualifying dominant inheritance context or documented cis/trans observation with a pathogenic variant is available.
BP4 Not met: REVEL was 0.313, above the generic missense BP4 threshold of <0.250 and within the 0.250–0.750 gray zone.
BP5 Not assessed: no alternate molecular basis for disease was documented in the available case evidence.
BP6 Not met: ClinVar reports p.Arg2127Gln as Uncertain significance, with zero expert-panel submissions and no exact benign classification.
N/A · 4 PVS1 · PM4 · BP3 · BP7
Research & evidence
Population frequency
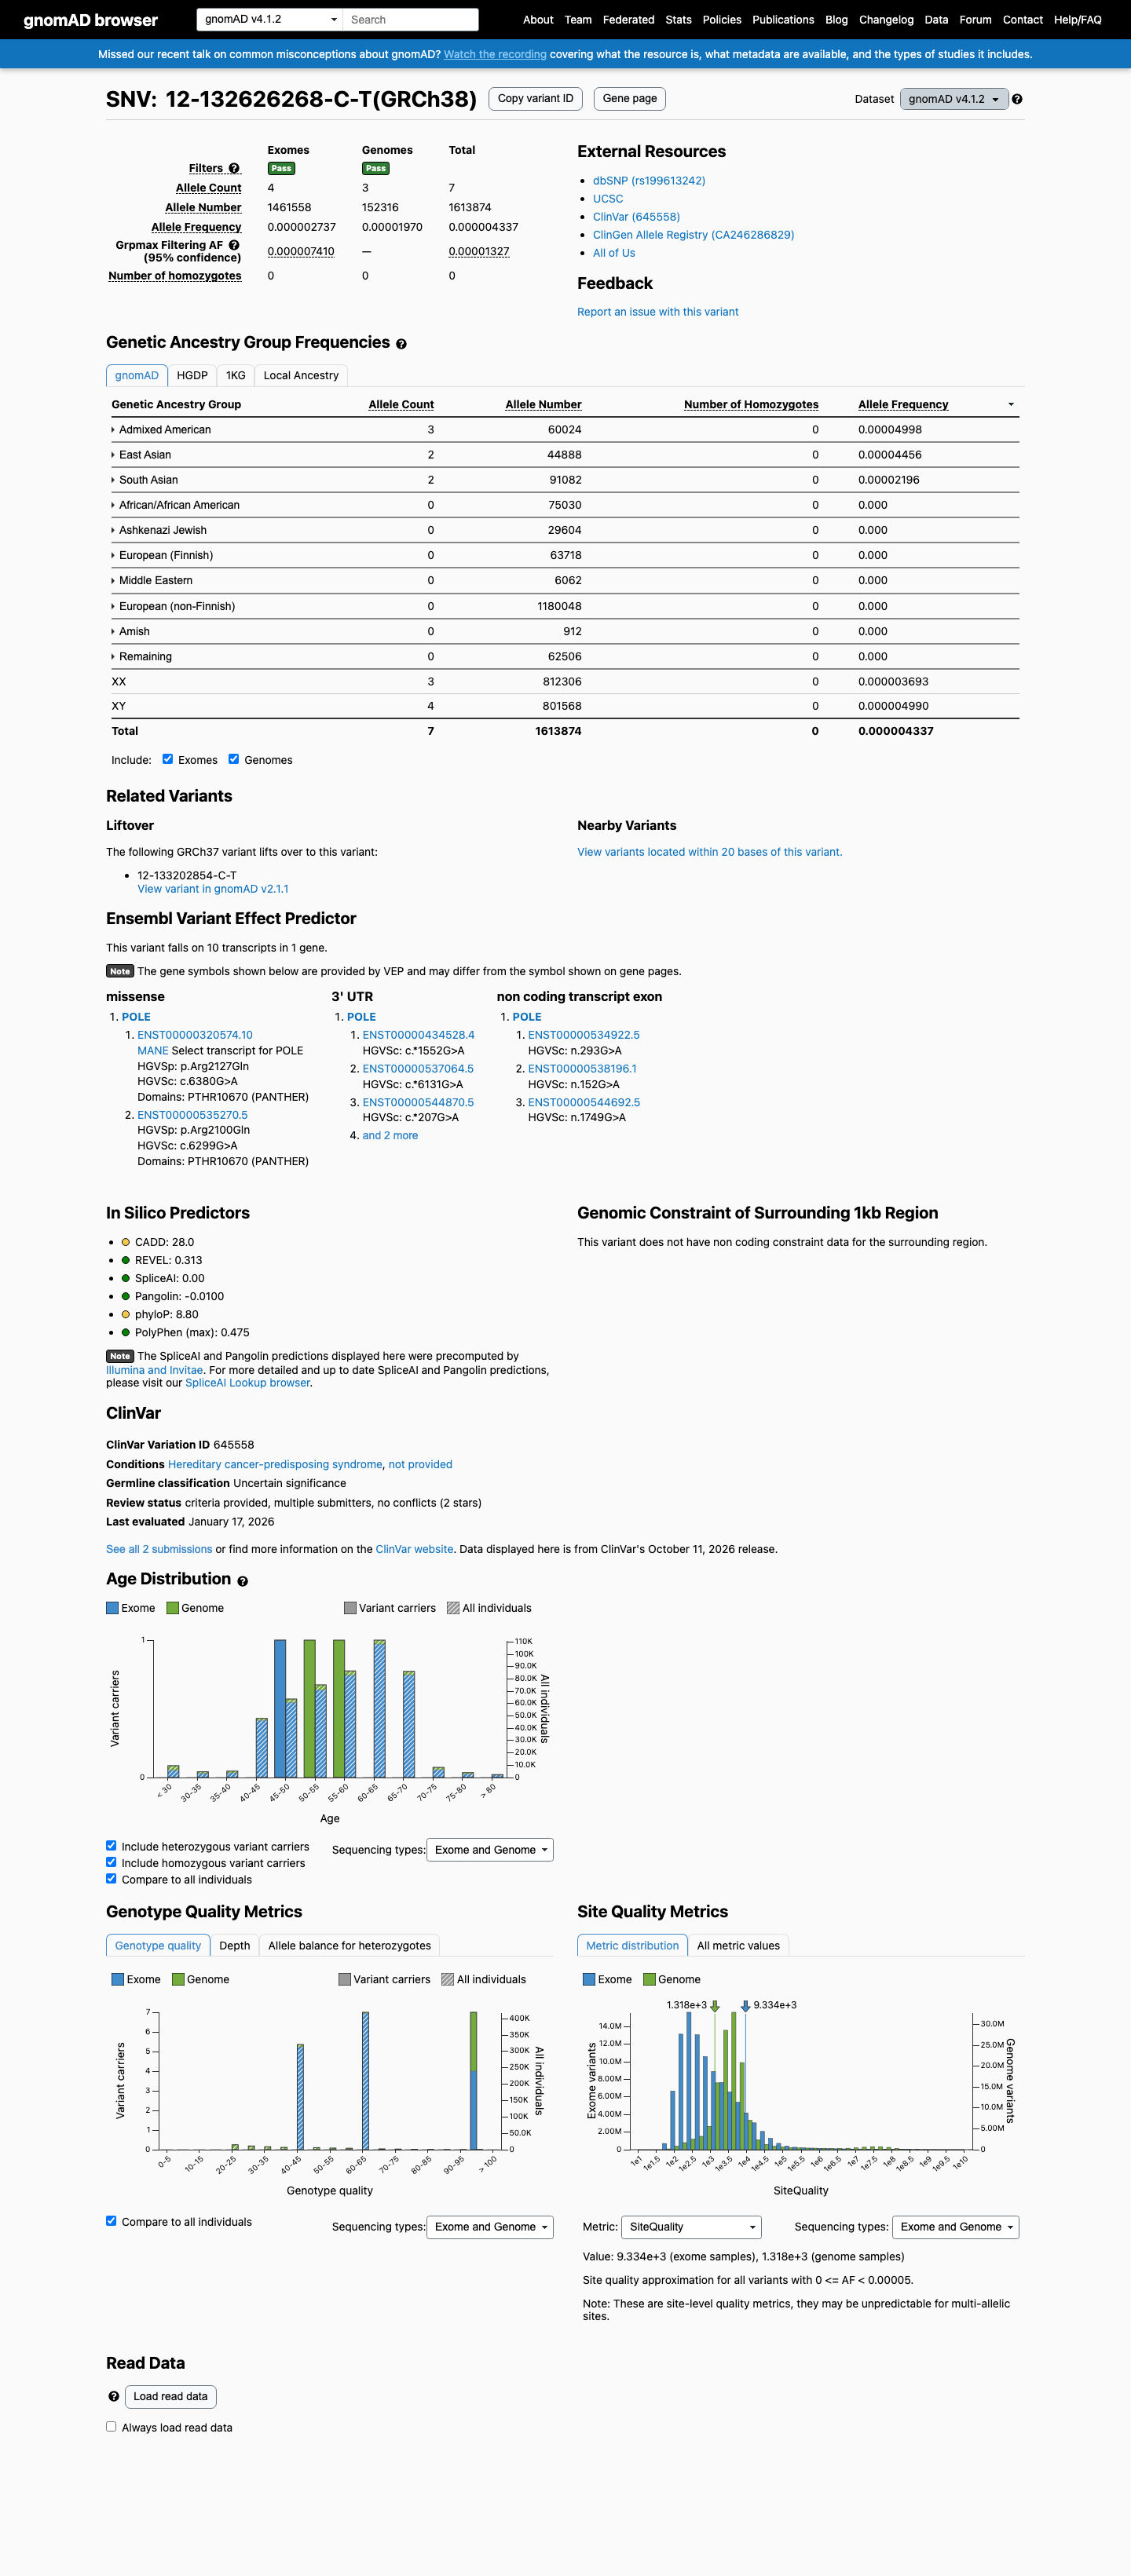
gnomAD v4.1 screenshot
gnomAD v4.1
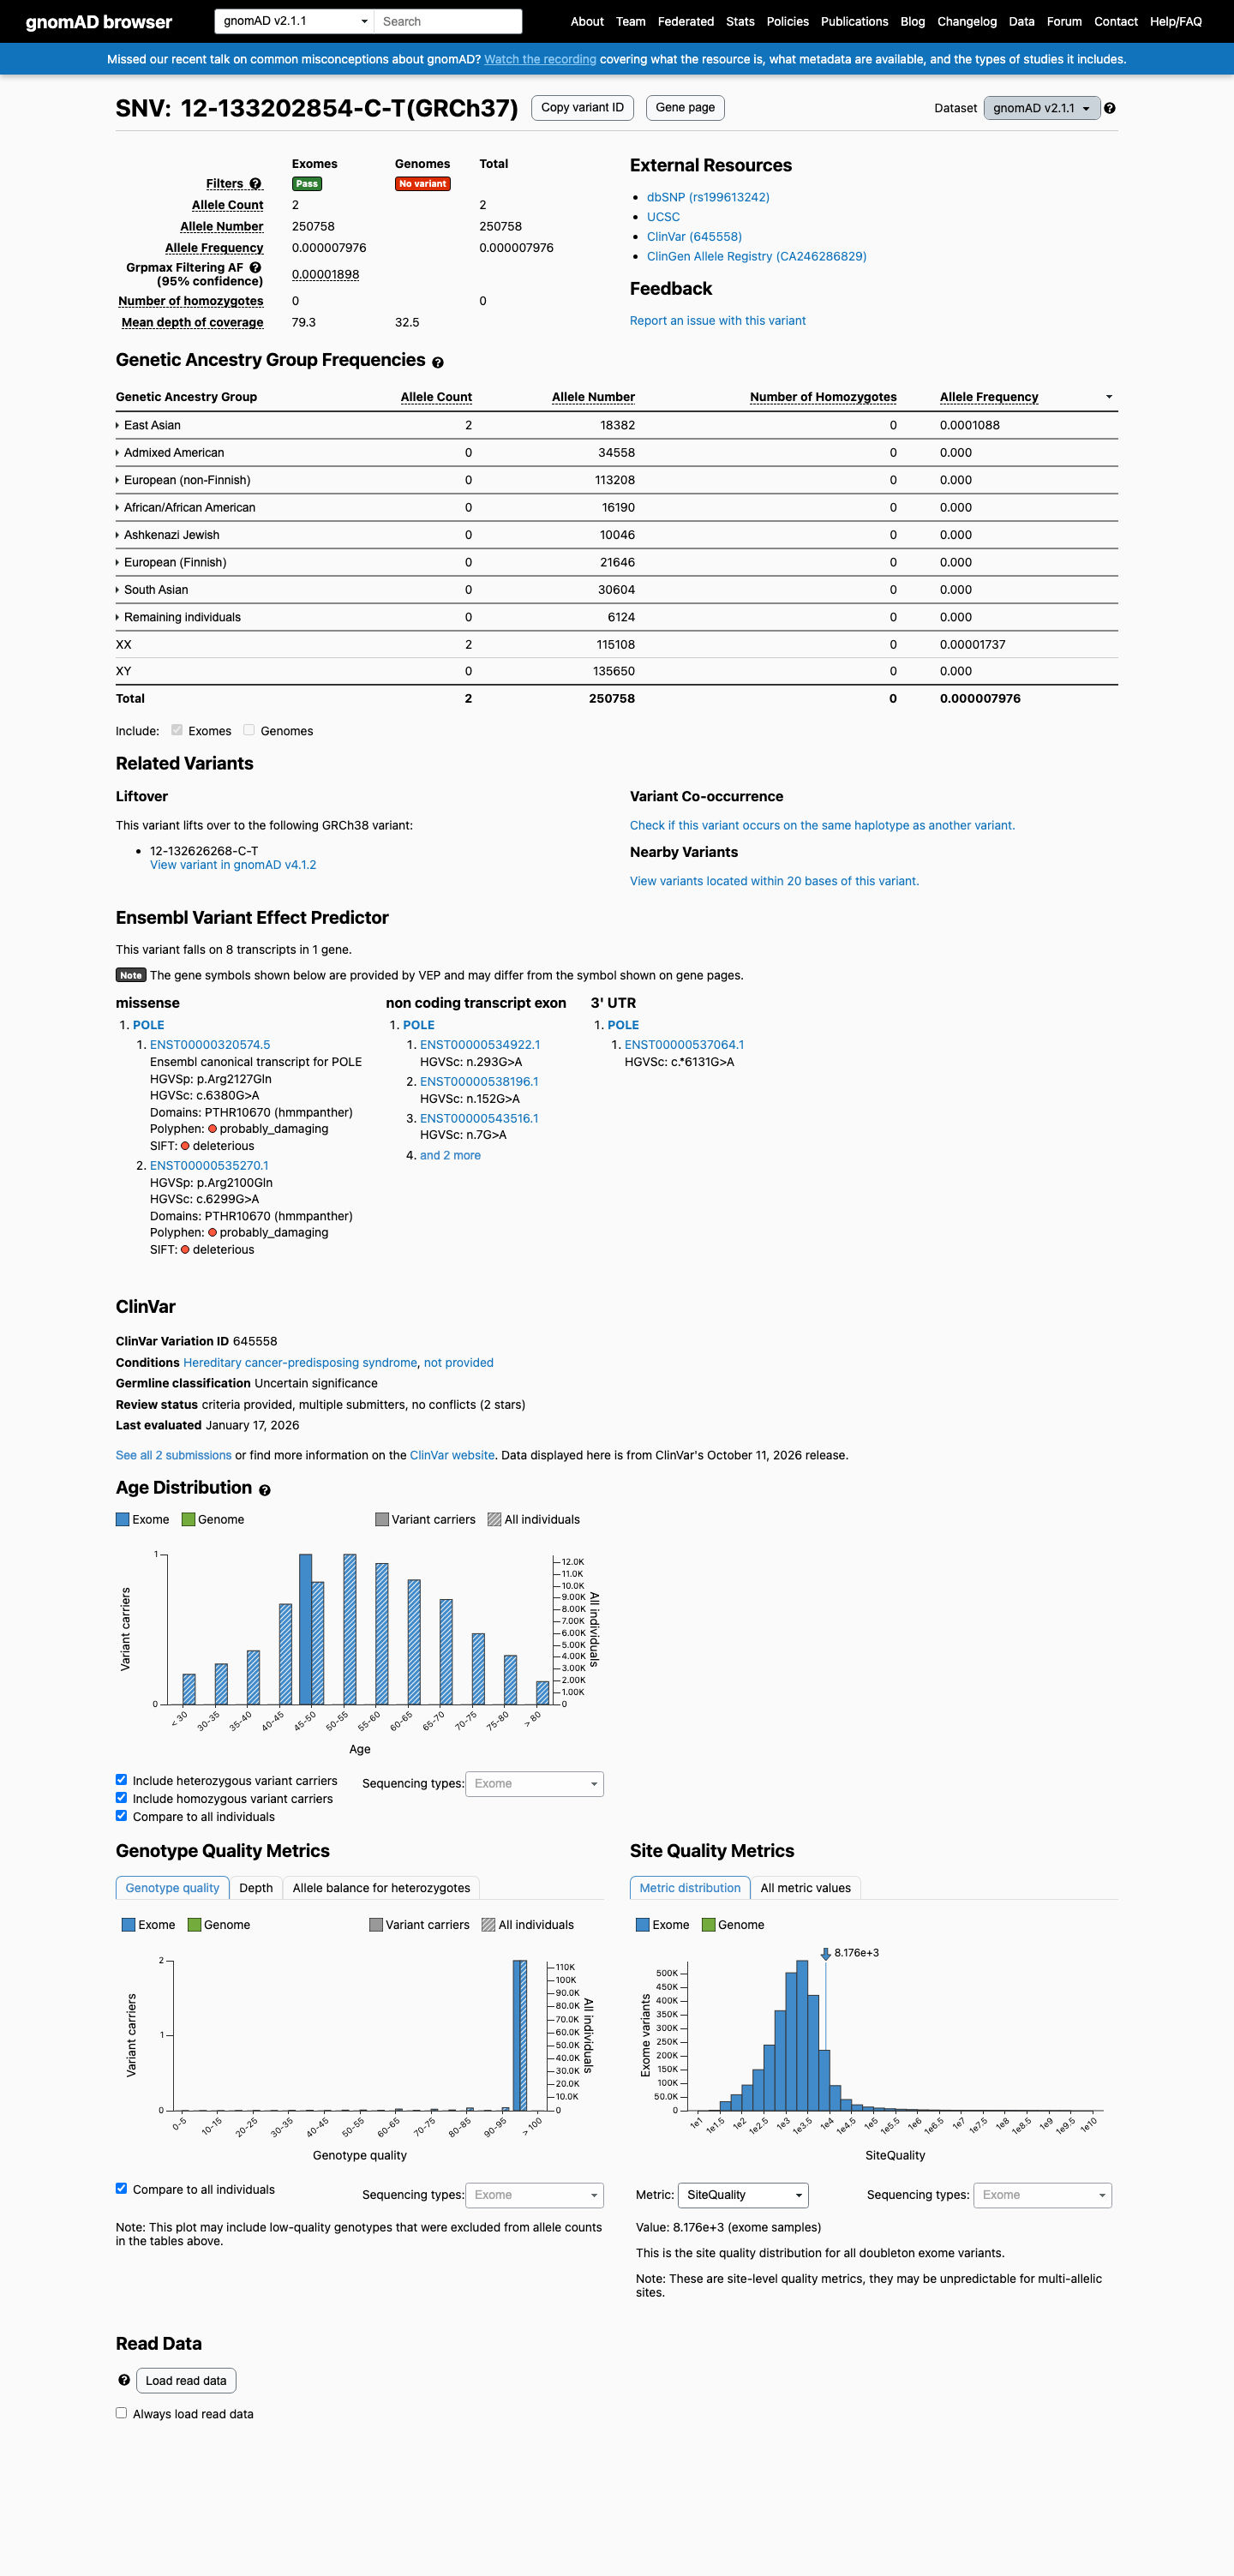
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 4.33739e-06; MAF= 0.00043%, 7/1613874 alleles, homozygotes = 0) and has highest observed frequency in the Admixed American population (AF= 4.998e-05; MAF= 0.00500%, 3/60024 alleles, homozygotes = 0); grpmax FAF= 1.327e-05.
v2.1
This variant is present in gnomAD v2.1 (AF= 7.97582e-06; MAF= 0.00080%, 2/250758 alleles, homozygotes = 0) and has highest observed frequency in the East Asian population (AF= 0.000108802; MAF= 0.01088%, 2/18382 alleles, homozygotes = 0); grpmax FAF= 1.898e-05.
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.00043% · 7 / 1,613,874
0 hom · FAF 0.0013%
Admixed American
3 / 60,024
0.005%
East Asian
2 / 44,888
0.0045%
South Asian
2 / 91,082
0.0022%
+ 7 not observed (Remaining individuals, European (Finnish), Amish, Middle Eastern, Ashkenazi Jewish, African/African American, European (non-Finnish))
gnomAD v2.1
0.0008% · 2 / 250,758
0 hom · FAF 0.0019%
East Asian
2 / 18,382
0.011%
+ 7 not observed (African/African American, Admixed American, Ashkenazi Jewish, European (Finnish), European (non-Finnish), Remaining individuals, South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
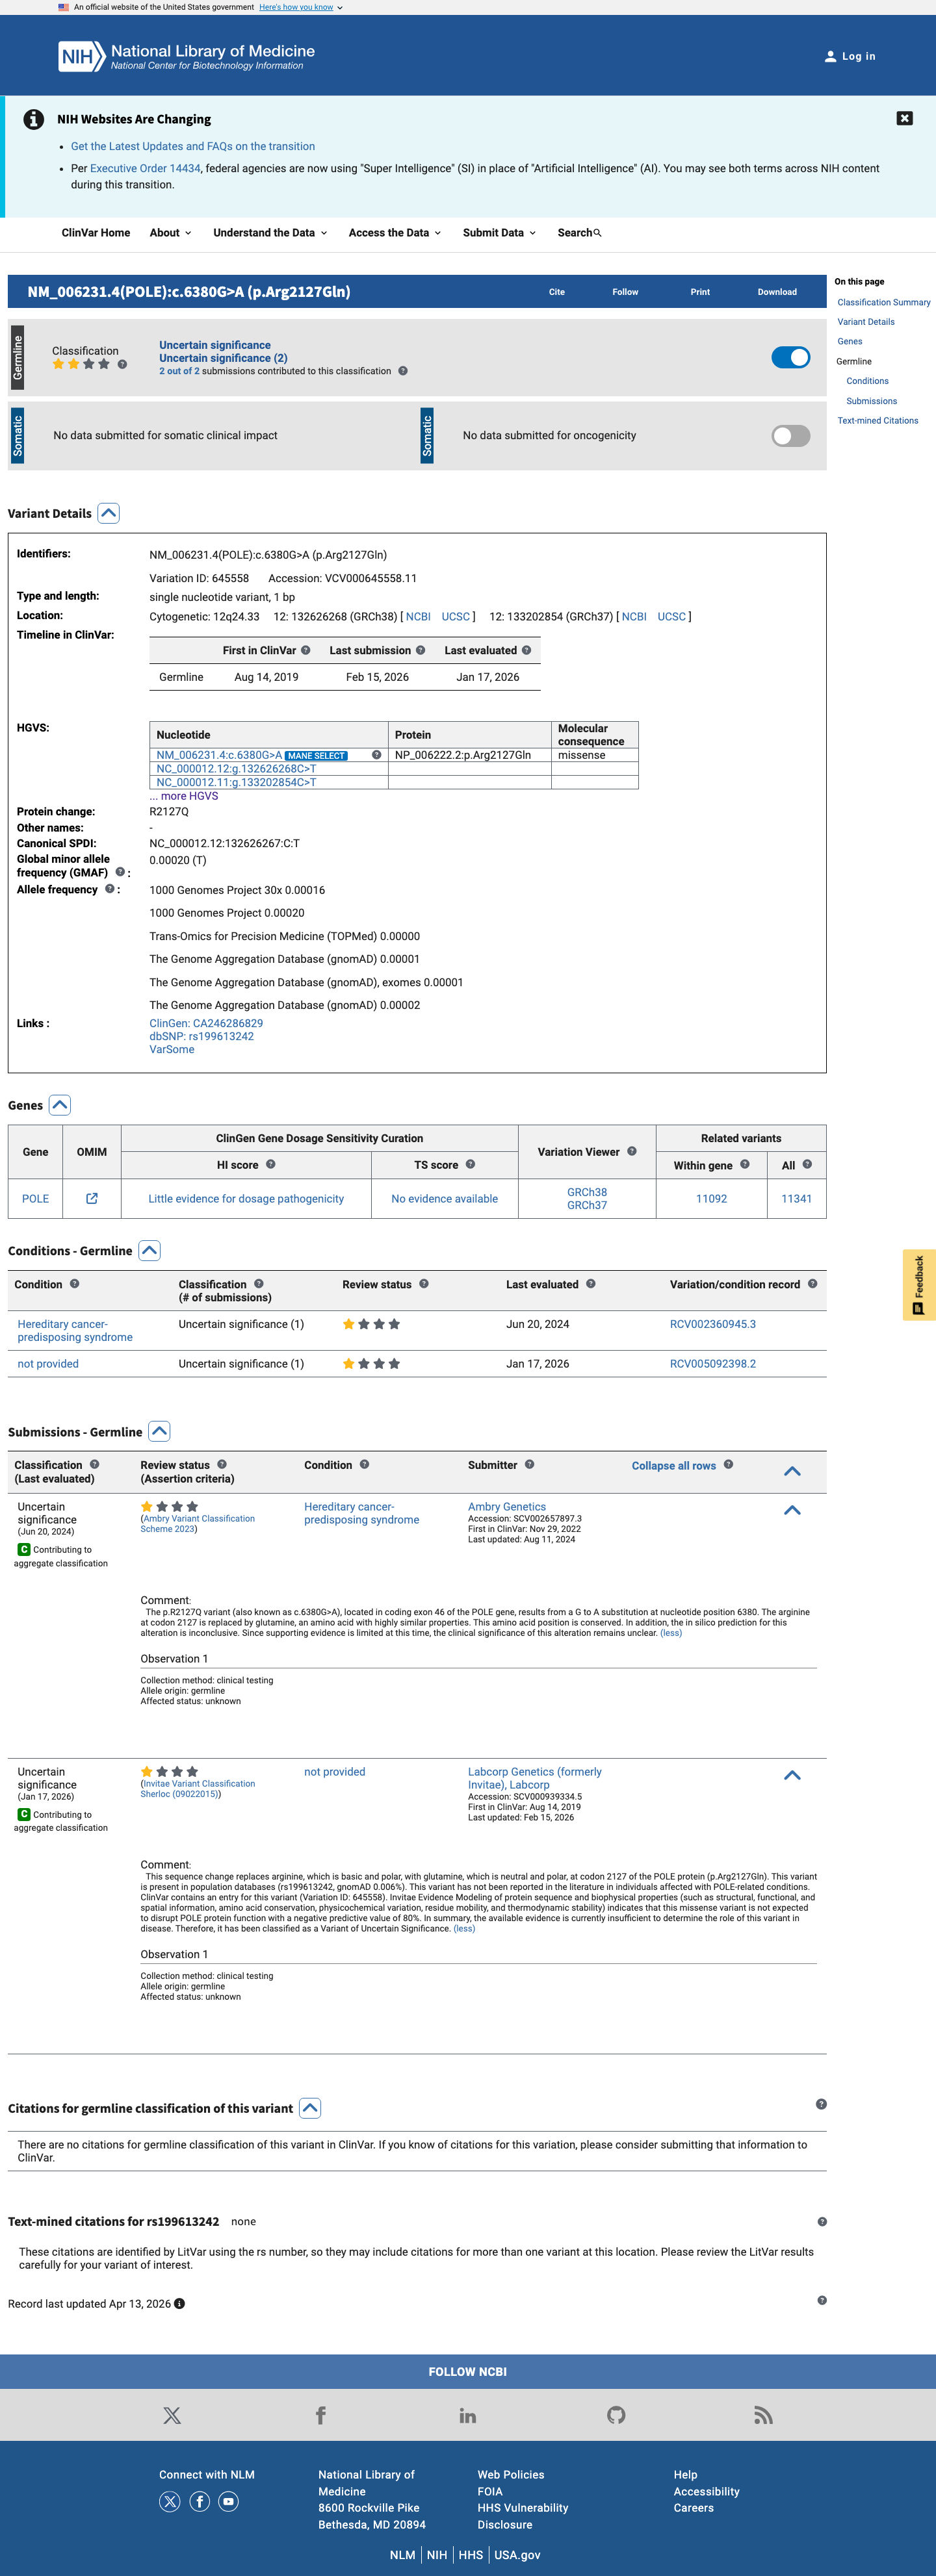
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Uncertain significance (2 clinical laboratories). (ClinVarID = 645558)
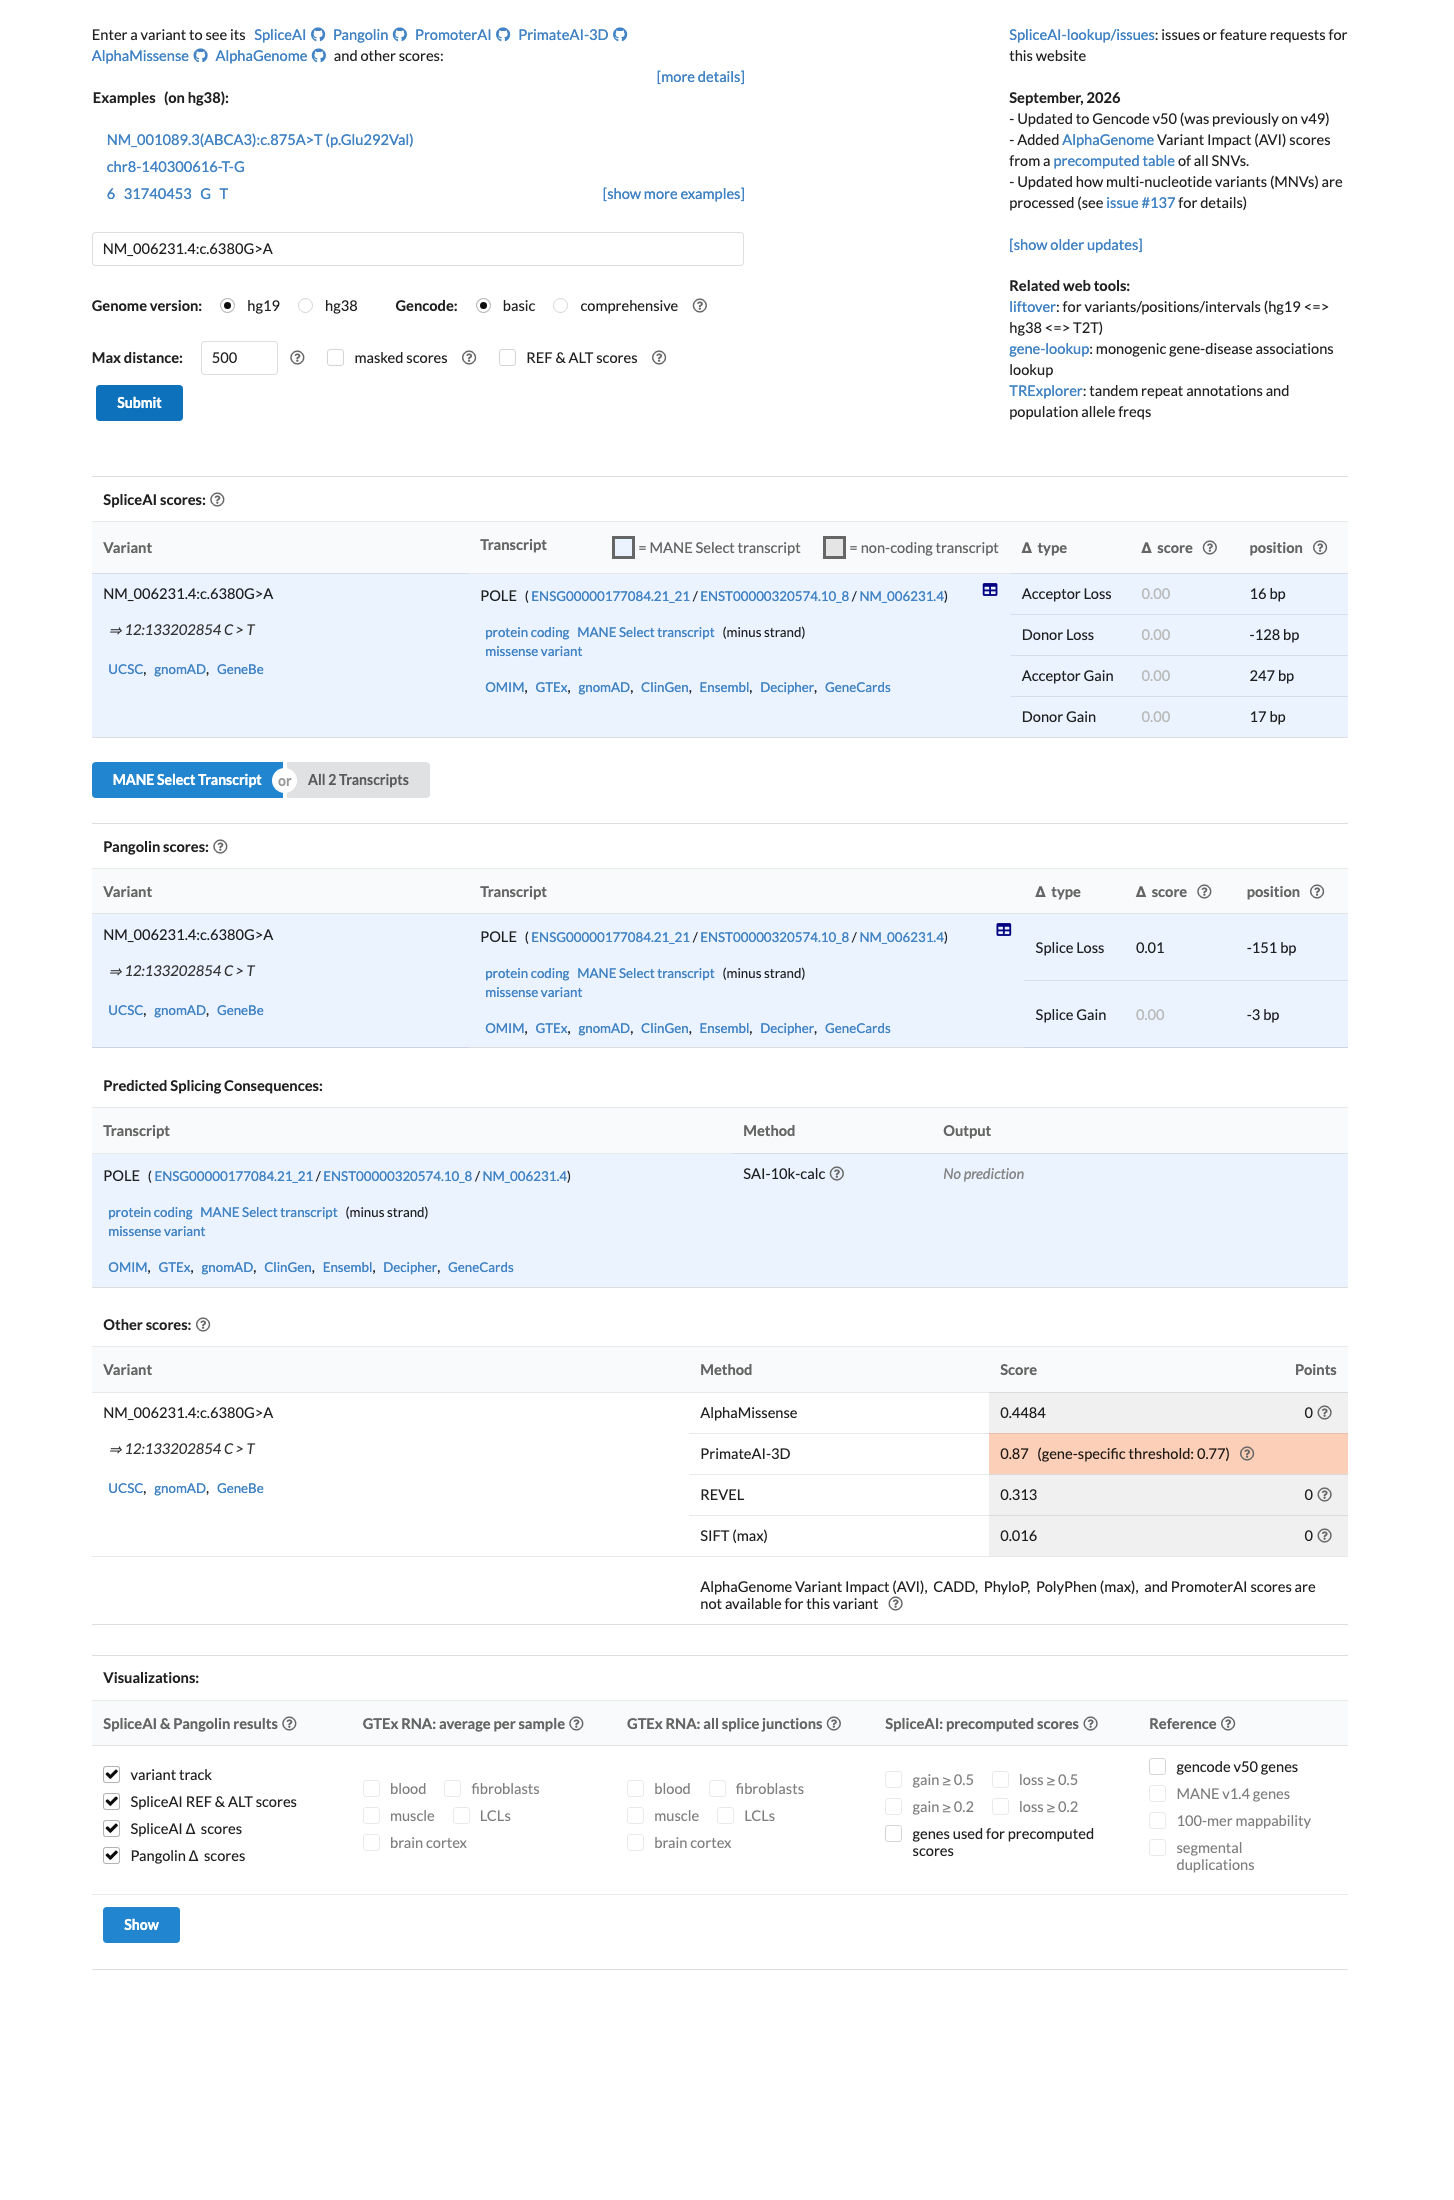
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.00). REVEL score = 0.313. BayesDel score = 0.0875751.
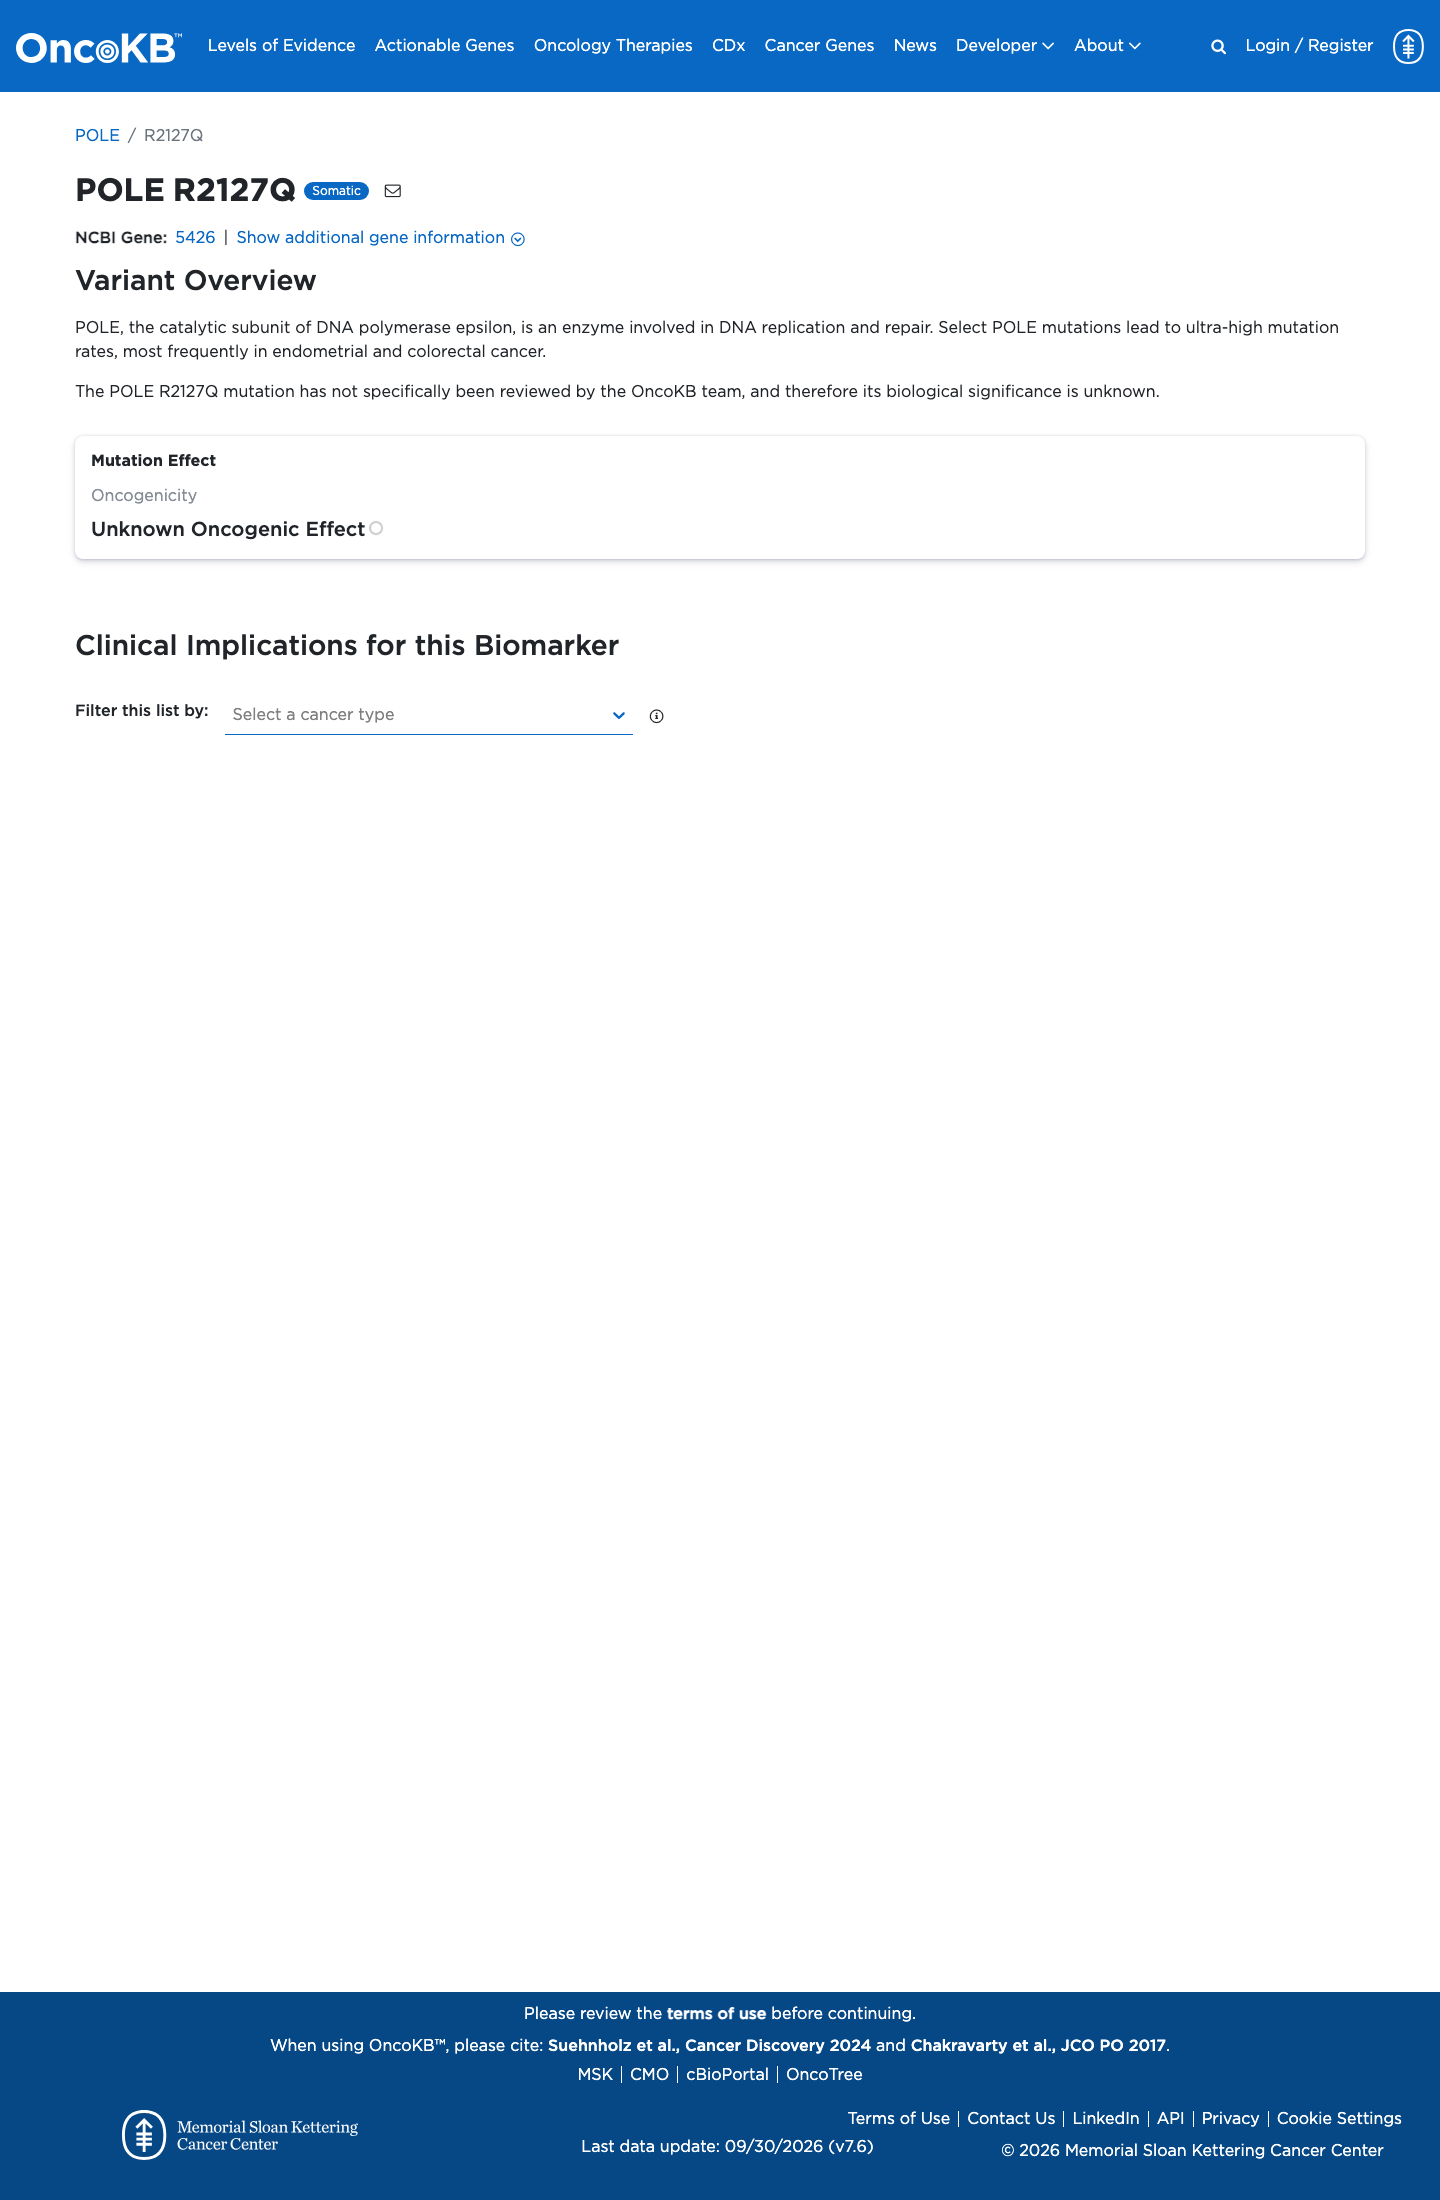
Functional / OncoKB screenshot
Functional Unknown Oncogenic Effect
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. POLE, the catalytic subunit of DNA polymerase epsilon, is an enzyme involved in DNA replication and repair. Select POLE mutations lead to ultra-high m
OncoKB ↗
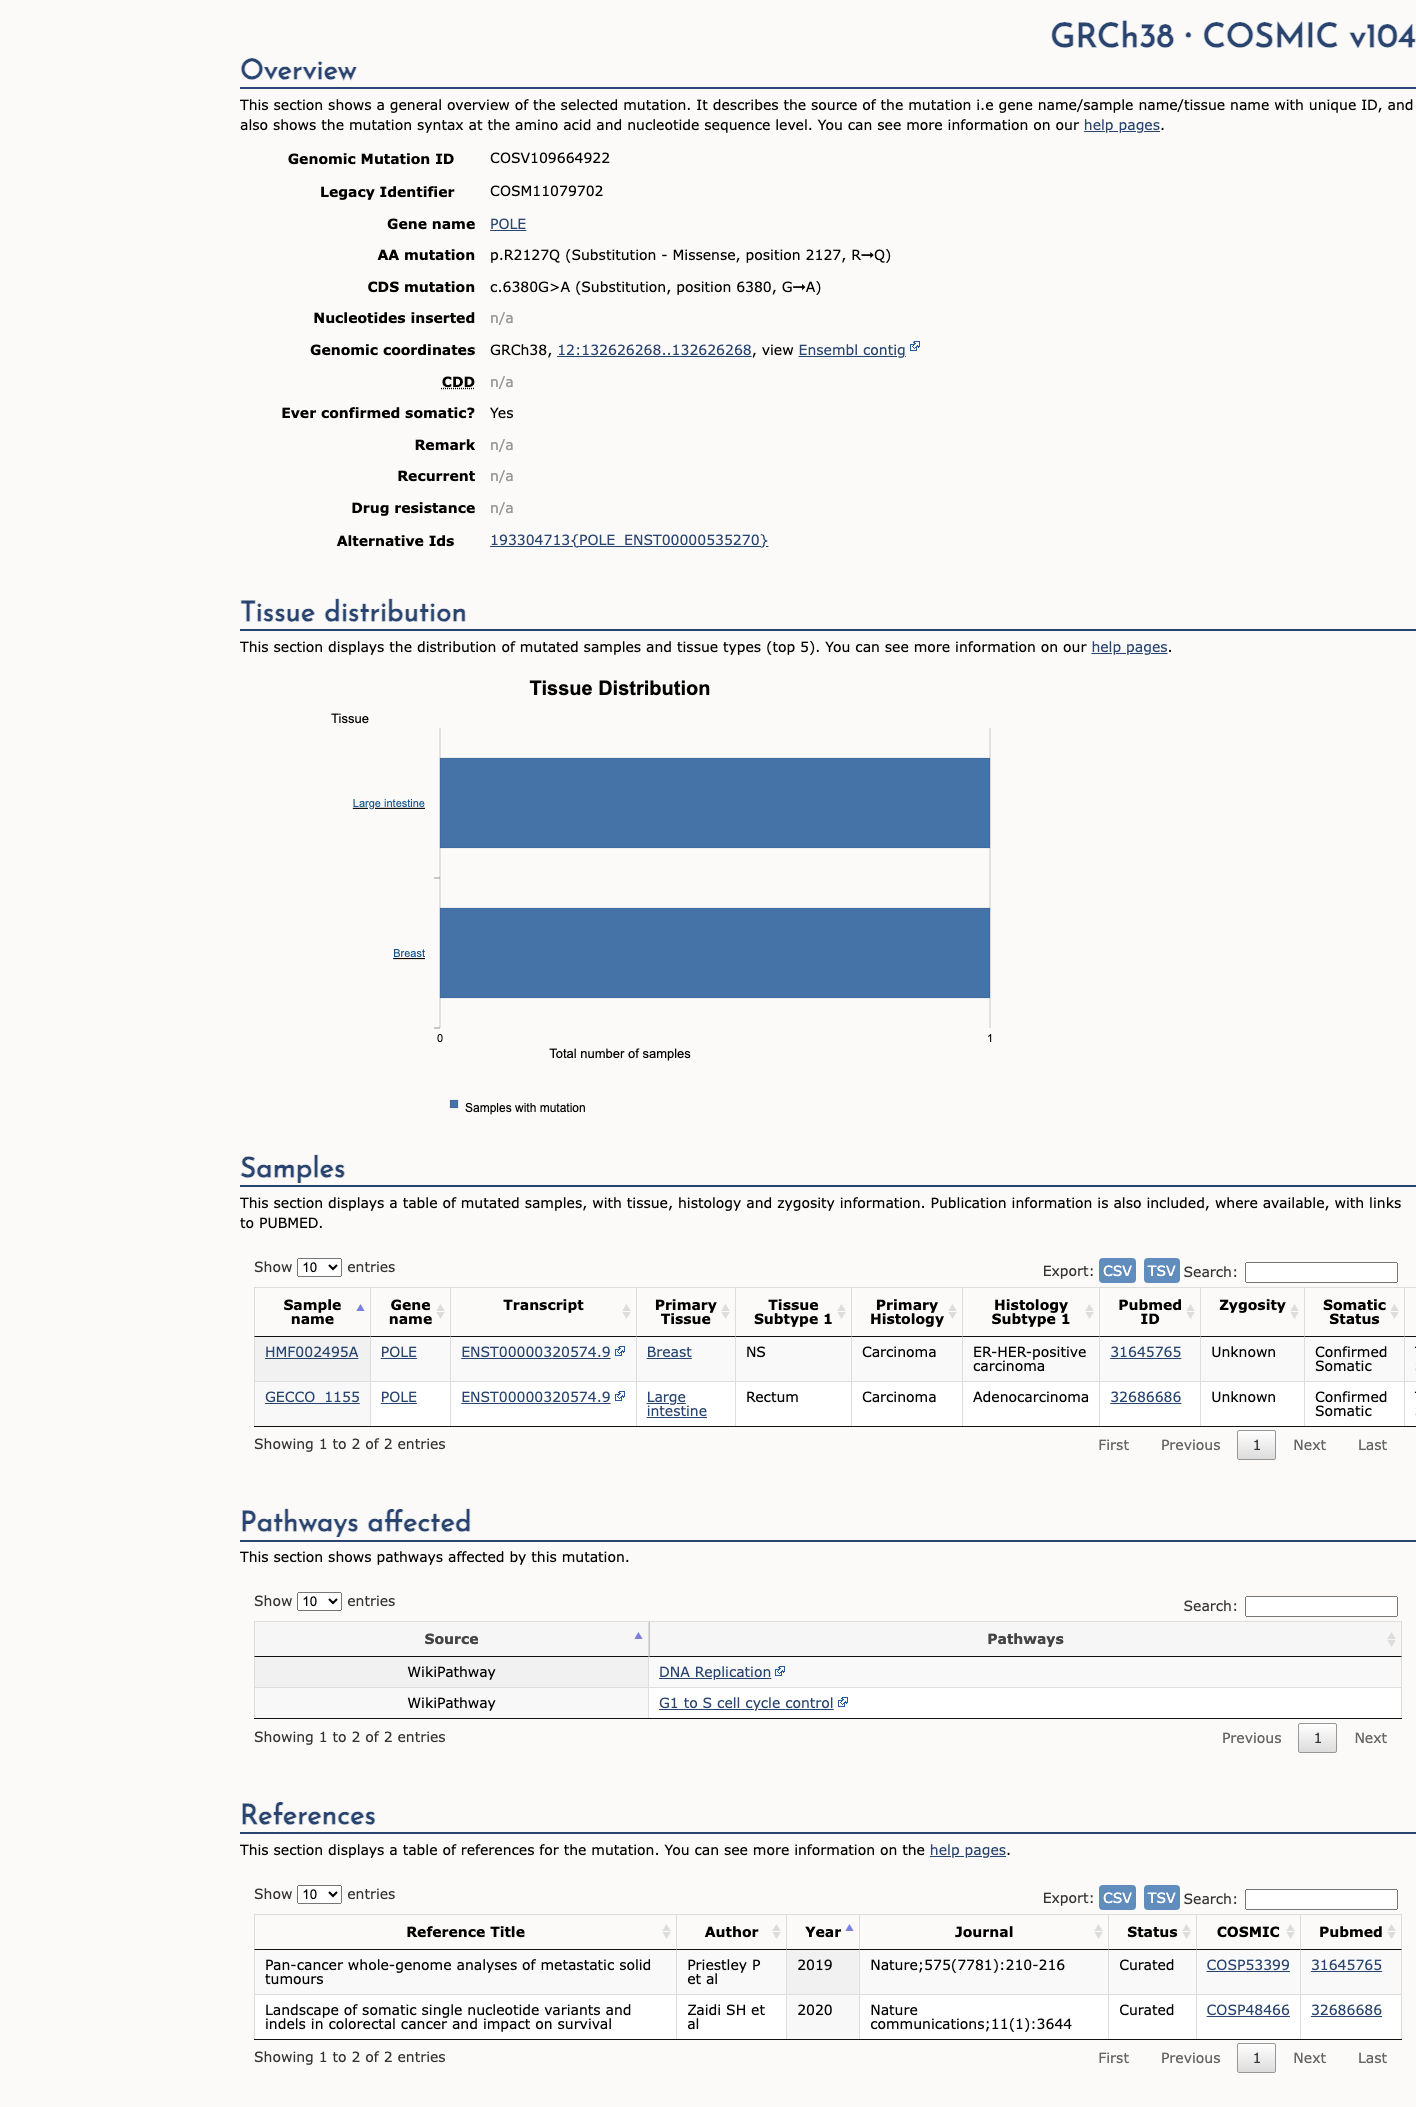
COSMIC screenshot
COSMIC
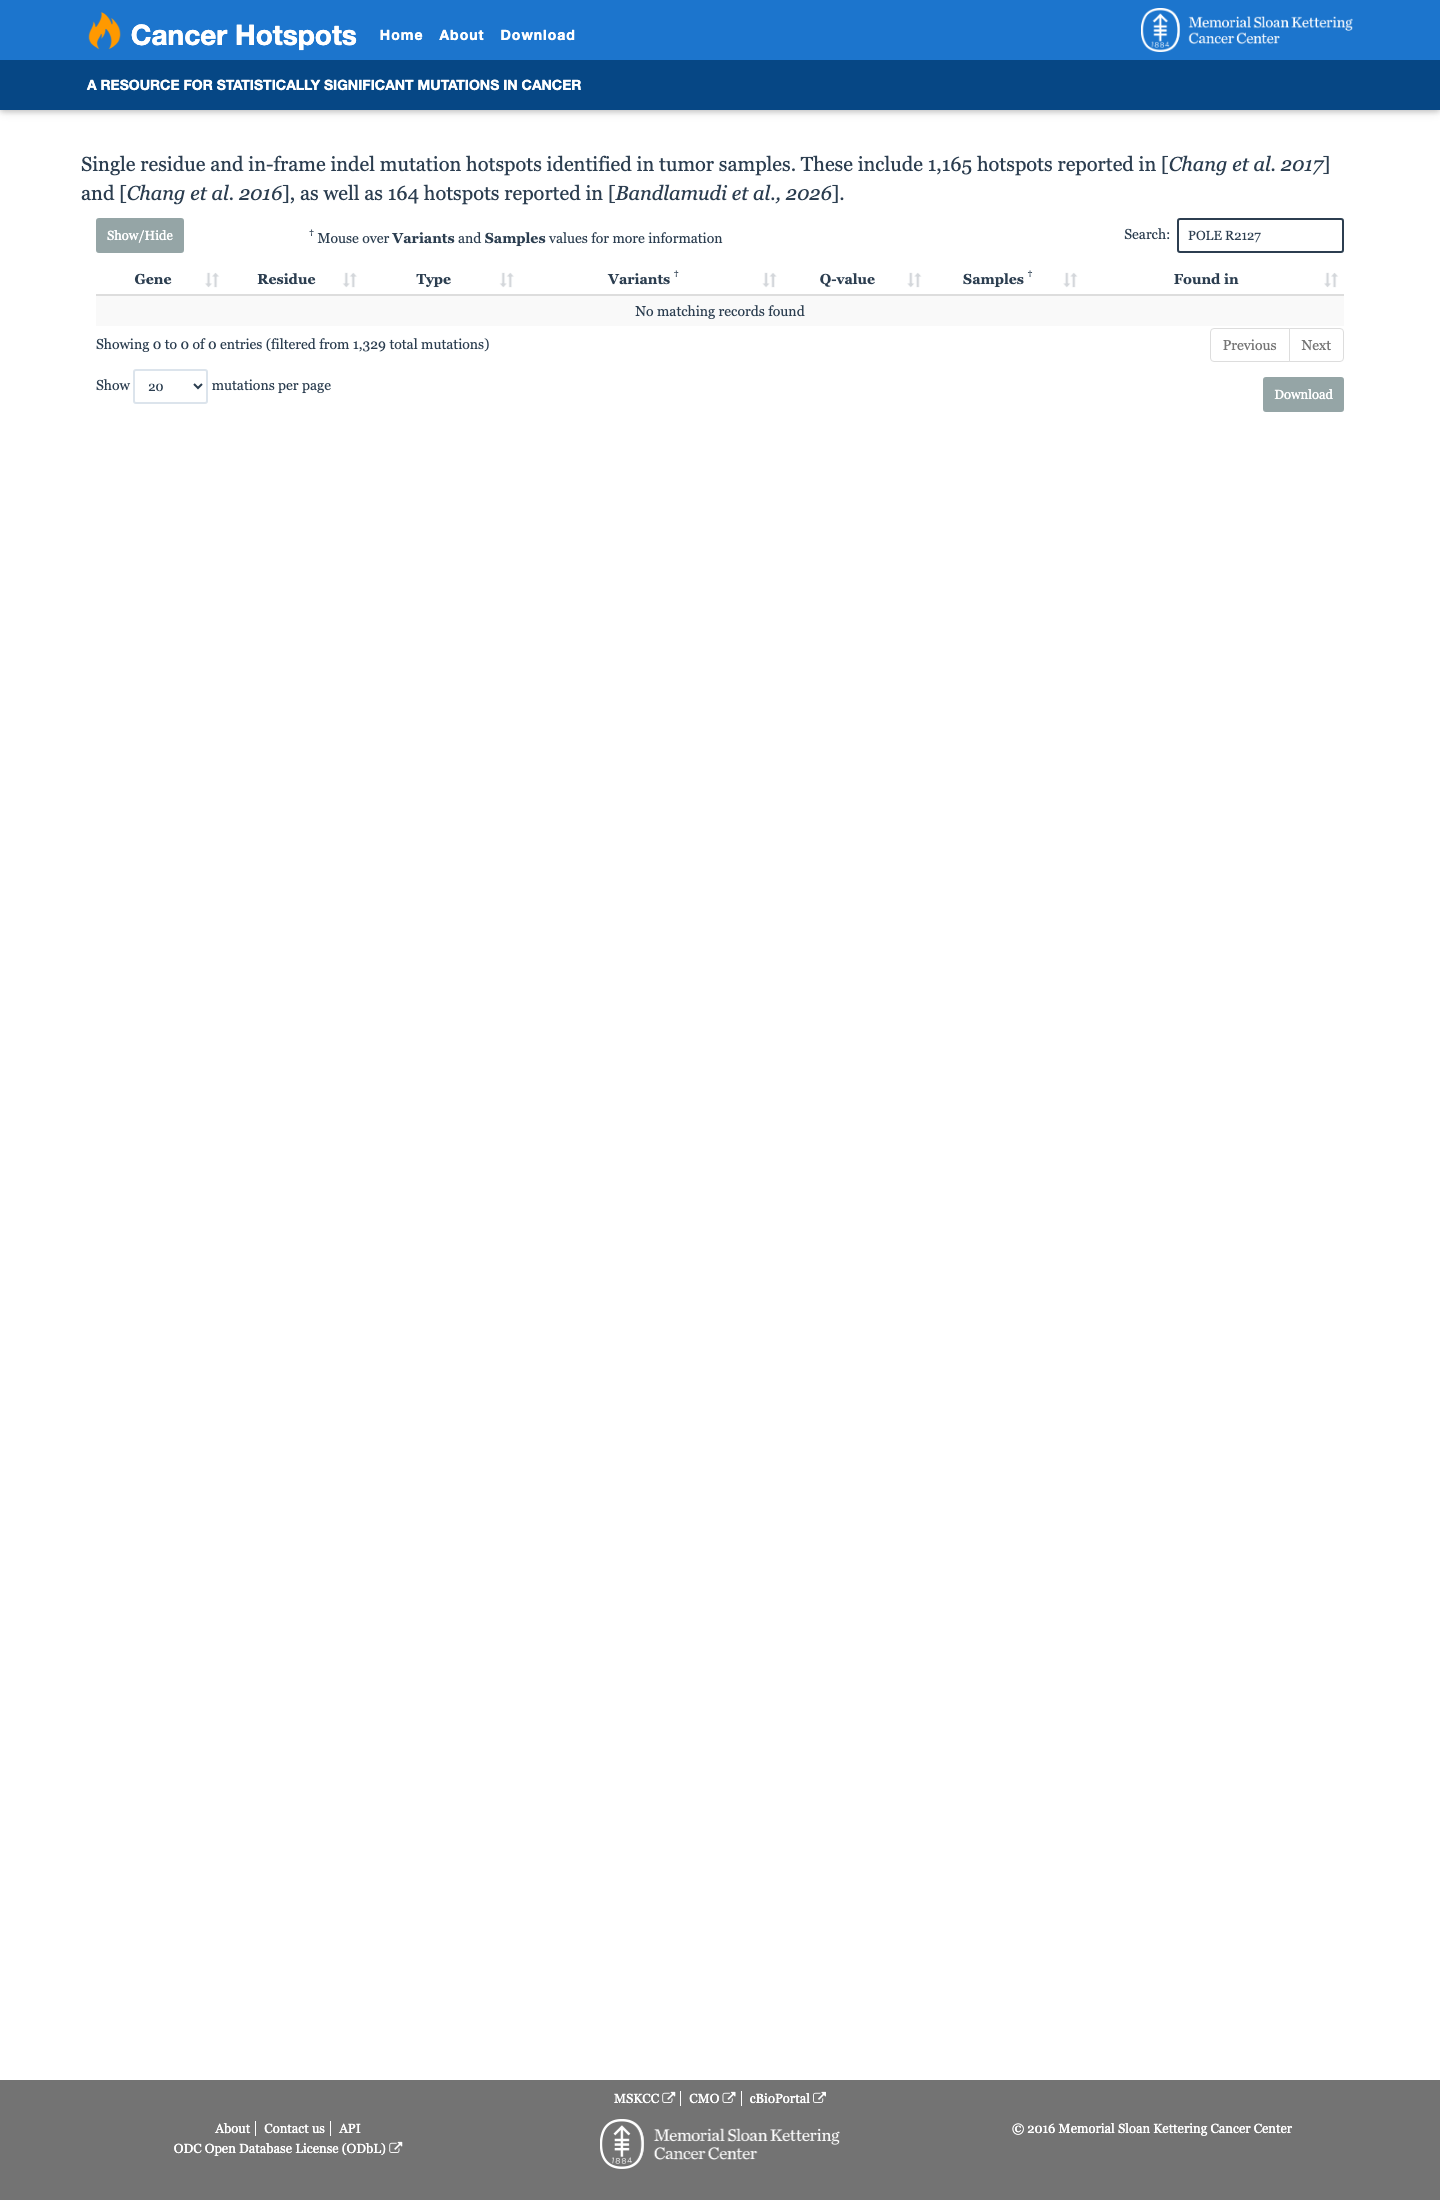
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has previously been reported in somatic cancers (COSMIC; COSV109664922, n = 2 times).
Hotspots
This variant does not lie in a statistically significant hotspot.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 1 PMID not cited in assessment
25394175 ↗ A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment. CLINVAR