ATM encodes a kinase that acts as a master controller of the cellular DNA damage response: it detects double-strand breaks and coordinates DNA repair, cell cycle arrest, or apoptosis through downstream targets including p53, BRCA1, and CHK2. Loss-of-function mutations in both copies of ATM cause ataxia telangiectasia, an autosomal recessive disorder characterized by neurological problems, immune deficiency, and cancer predisposition. ATM functions as a tumor suppressor: individuals with ataxia telangiectasia are prone to childhood lymphomas, leukemias, and breast cancer, while carriers of a single altered copy have increased risk of breast, pancreatic, prostate, and other cancers. Somatic ATM mutations also occur in lymphoid malignancies and solid tumors, and ATM-deficient cancers are often especially sensitive to DNA-damaging treatments.
This variant
As a loss-of-function change in ATM, a tumor-suppressor gene that coordinates DNA-double-strand-break repair, this variant is relevant to ATM-related cancer predisposition and autosomal-recessive ataxia-telangiectasia.
Transcript
NM_000051.4
HGVS · transcript:coding
NM_000051.4:c.3077+1G>A
GRCh38
chr11:108271407 G>A
GRCh37
chr11:108142134 G>A
Likely Pathogenic: ATM VCEP Rule12 is met by PVS1 strong plus PS3 and PM2 supporting.
Classification rationale
PVS1PS3PM2Likely Pathogenic
ATM c.3077+1G>Acanonical_splice · exon 20i
Likely Pathogenic: PVS1 strong reflects a canonical +1 splice disruption with an in-frame exon-20 skip and an intact protein remainder. Likely Pathogenic: PS3 supporting reflects a high-confidence direct prime-editing result showing the exact variant is non-functional. Likely Pathogenic: PM2 supporting reflects a gnomAD maximum subpopulation frequency below the ATM VCEP rarity threshold.
PVS1 + PS3 + PM2→Likely Pathogenic
LYFE Sciences is an AI system, and it can make mistakes. Criteria
may be applied incorrectly, sources may be misread, and a confident-looking
classification can still be wrong. Double-check every criterion and
its underlying evidence before relying on any call.
Gene diagram
· NM_000051.4 · variants mapped to exon structure
ATMNM_000051.4
Fetching transcript structure from UCSC…
Exons
—
Transcript span
—
Strand
—
Variants mapped
—
Source
UCSC ncbiRefSeqCurated
All variants in ATM—click a row to locate it on the plot · use the link column to open its page
Protein
Location
Classification
Link
Applied criteria · 3 applied · 9 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 3
Strength Supporting Moderate Strong Very strong
✓
PVS1strongPathogenic
Met, strong: canonical +1 splice disruption predicts an in-frame 156-nucleotide exon-20 skip, triggering the VCEP's intact-remainder/escape-from-NMD downgrade rule.
Variant assessment: NM_000051.4:c.3077+1G>A; NP_000042.3:p.?; canonical donor (+1) splice variant; internal coding exon 20 of 63; exon 20 spans c.2922_3077 and contains 156 coding nucleotides.Variant assessment: predicted exon-20 skipping removes 156 coding nucleotides, with 156 % 3 = 0 and preserved reading frame; cryptic splice-site use is not modeled.ATM_PVS1_1.6.pdf, page 1: canonical +/-1 or 2 splice sites are PVS1-eligible null variants when LOF is a known disease mechanism; the same page cautions about exon skipping that leaves the remainder of the protein intact.
Met at Supporting: direct prime-editing row classifies c.3077+1G>A as Non-functional with High confidence and DeepATM_predicted=No.
Sun et al. 2025, Supplementary Table S1, row c.3077+1G>A: hg38 position 108271407, G>A, exon_20, Splice_AD, Classification=Non-functional, Confidence=High, DeepATM_predicted=No, Combined_score=-3.24742909184742, ClinVar_classification=Likely pathogenic.The VCEP file-specific policy states that DeepATM_predicted=No with High, Medium-high, or Medium confidence represents a direct prime-editing measurement and that Classification=Non-functional supports PS3_Supporting.The approved functional assay table describes ATM kinase-activity and radiosensitivity methodology and strength calibration, but its variant-specific validation/result fields list other variants and do not contain c.3077+1G>A or p.? for this case.
Met at Supporting: highest observed gnomAD subpopulation AF is 0.0000847%, below the VCEP PM2 threshold of <=0.001%.
CSPEC criteria entry PM2, Supporting rule: Frequency <=.001% in gnomAD subpopulation with the highest frequency; singleton exception also specified.gnomAD v4.1 all-comers entry for NM_000051.4:c.3077+1G>A: highest observed subpopulation AF 8.474834825469252e-07 (0.0000847483%) in European (non-Finnish) individuals, 1/1,179,964 alleles, homozygotes 0.gnomAD v2.1 all-comers exomes entry: total AF 0 (0/250,488 alleles), highest observed subpopulation AF 0, homozygotes 0.
Assessed · not applied
· 4 not met · 5 not assessed
Pathogenic
PS1Not assessed: the governing ATM PS1 splicing table contains no c.3077+1G>A or p.? entry establishing a matching reference splice event.
PS4Not assessed: the only case-control result was gene-level ATM data (OR=1.88, lower 95% CI=1.10), not an exact-variant estimate.
PM3Not assessed: no qualifying affected-proband observation or phase-established pathogenic ATM variant in trans is available for PM3 point assignment.
PP1Not assessed: no documented affected-relative segregation count or variant-specific parental testing is available to meet the PP1 thresholds.
Benign
BA1Not met: highest observed gnomAD subpopulation AF is 8.47e-7, far below the VCEP BA1 threshold of >0.5%.
BS1Not met: highest observed gnomAD subpopulation AF is 0.0000847%, below the VCEP BS1 threshold of >0.05%.
BS3Not met: direct prime-editing measurement classifies c.3077+1G>A as Non-functional, not as rescuing an ATM-specific feature or radiosensitivity.
BP2Not assessed: no qualifying unaffected-adult observation documents this variant in trans or cis with a pathogenic ATM variant for BP2.
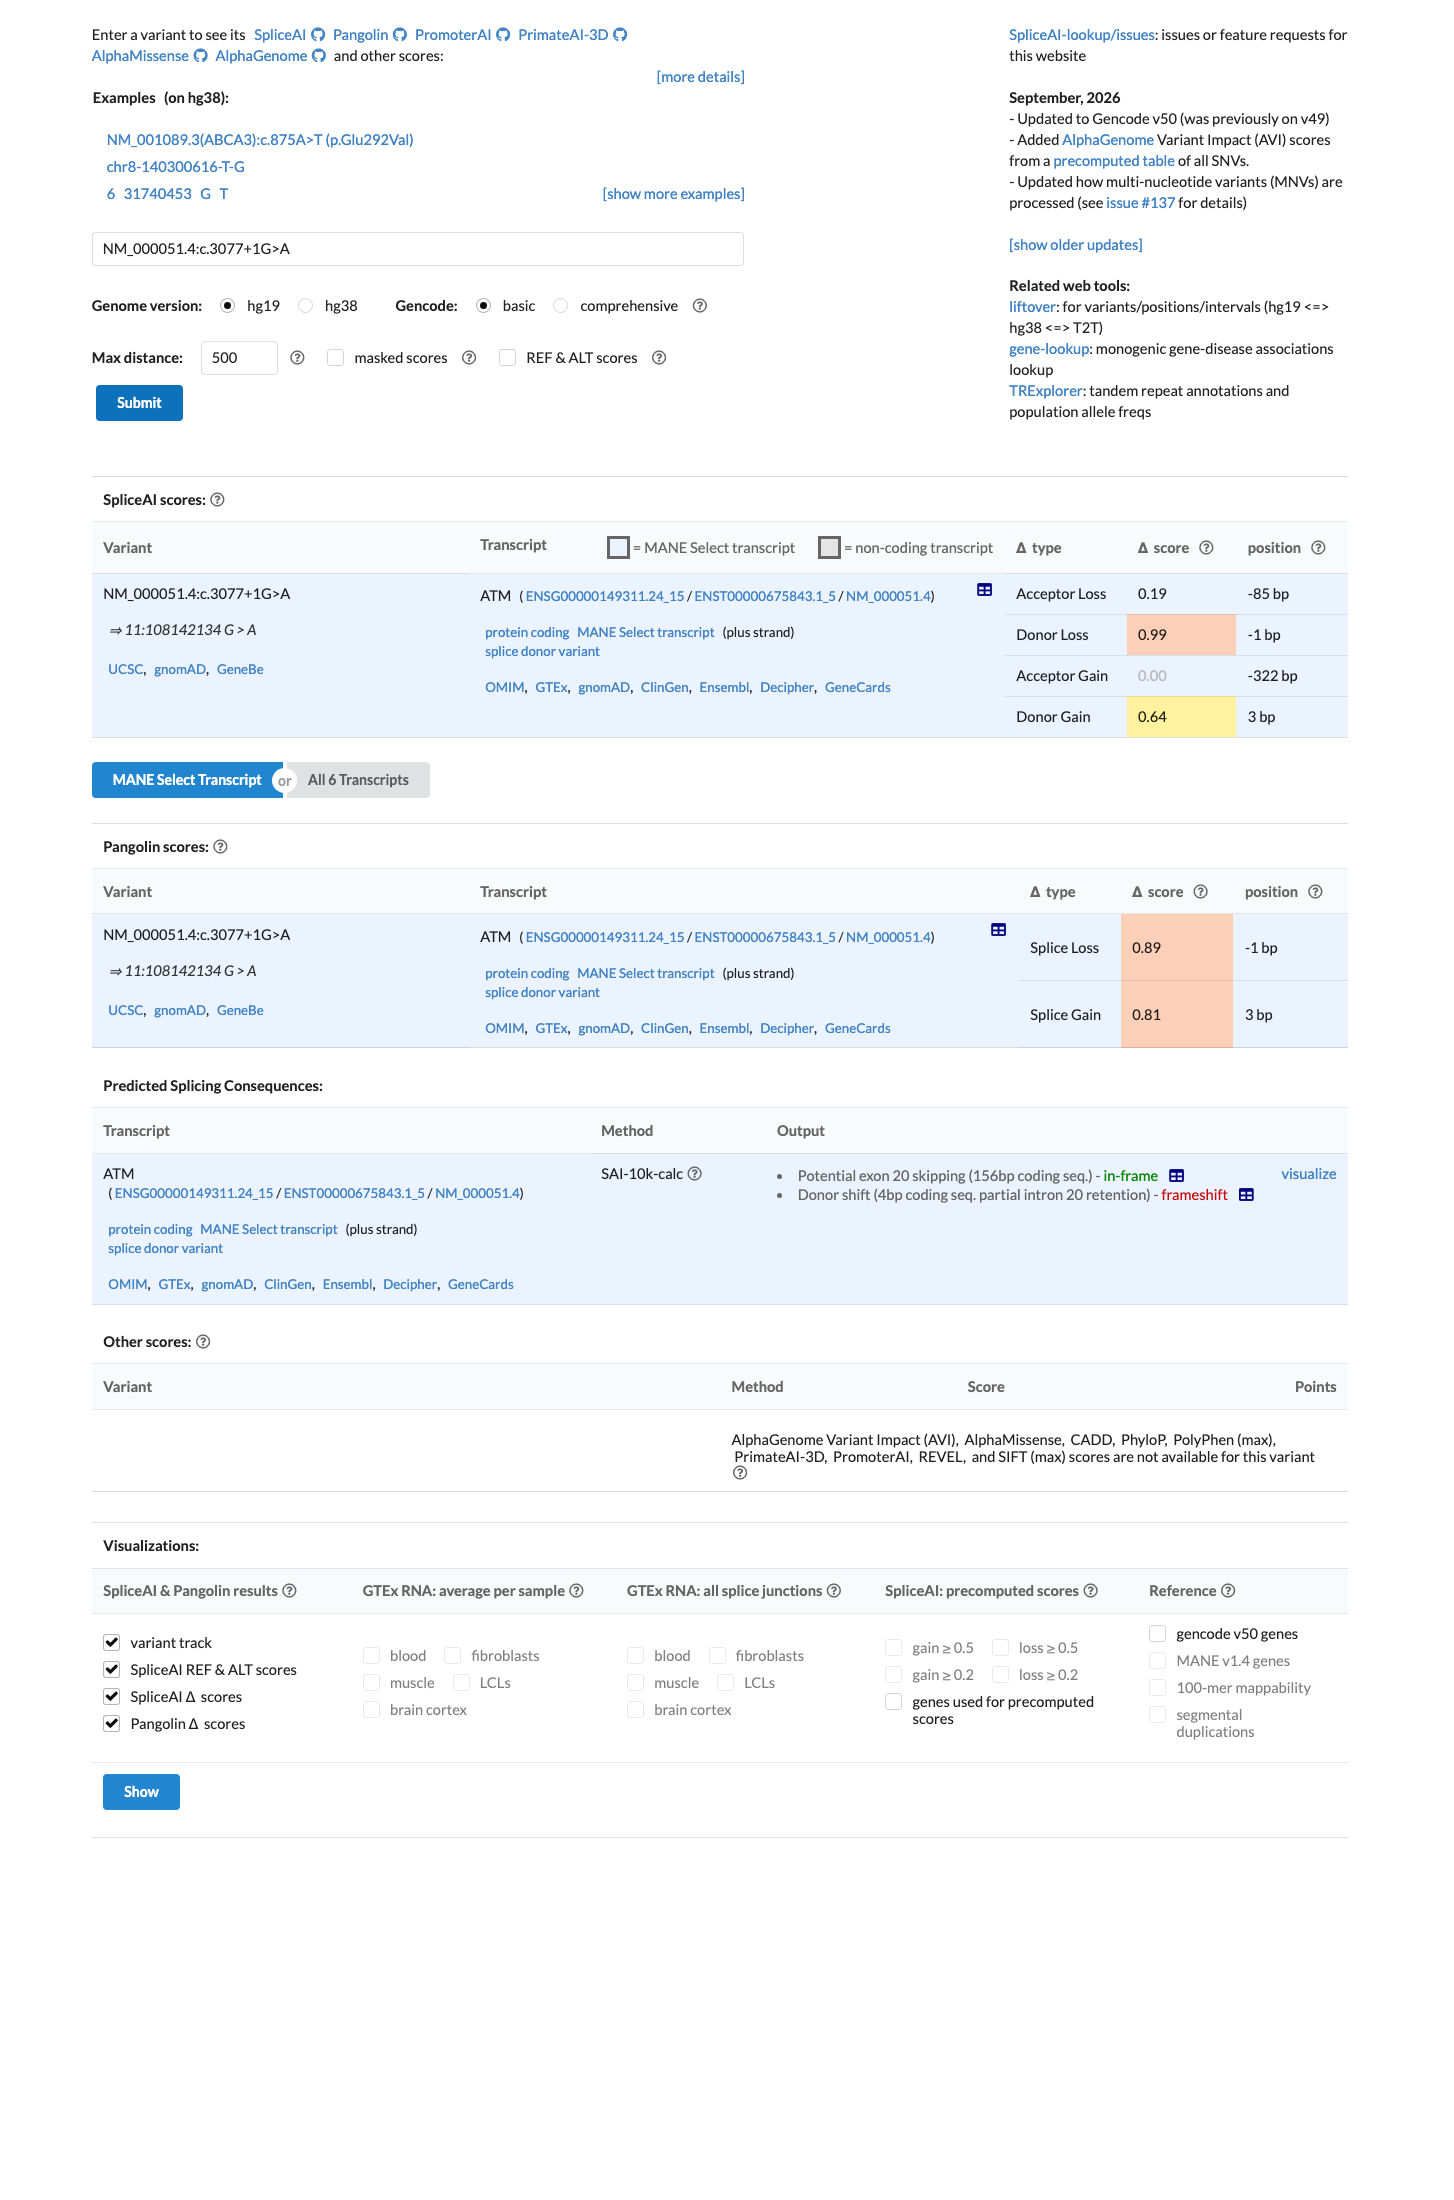
BP4Not met: SpliceAI max delta 0.992 exceeds the VCEP BP4 no-impact threshold of ≤0.1.
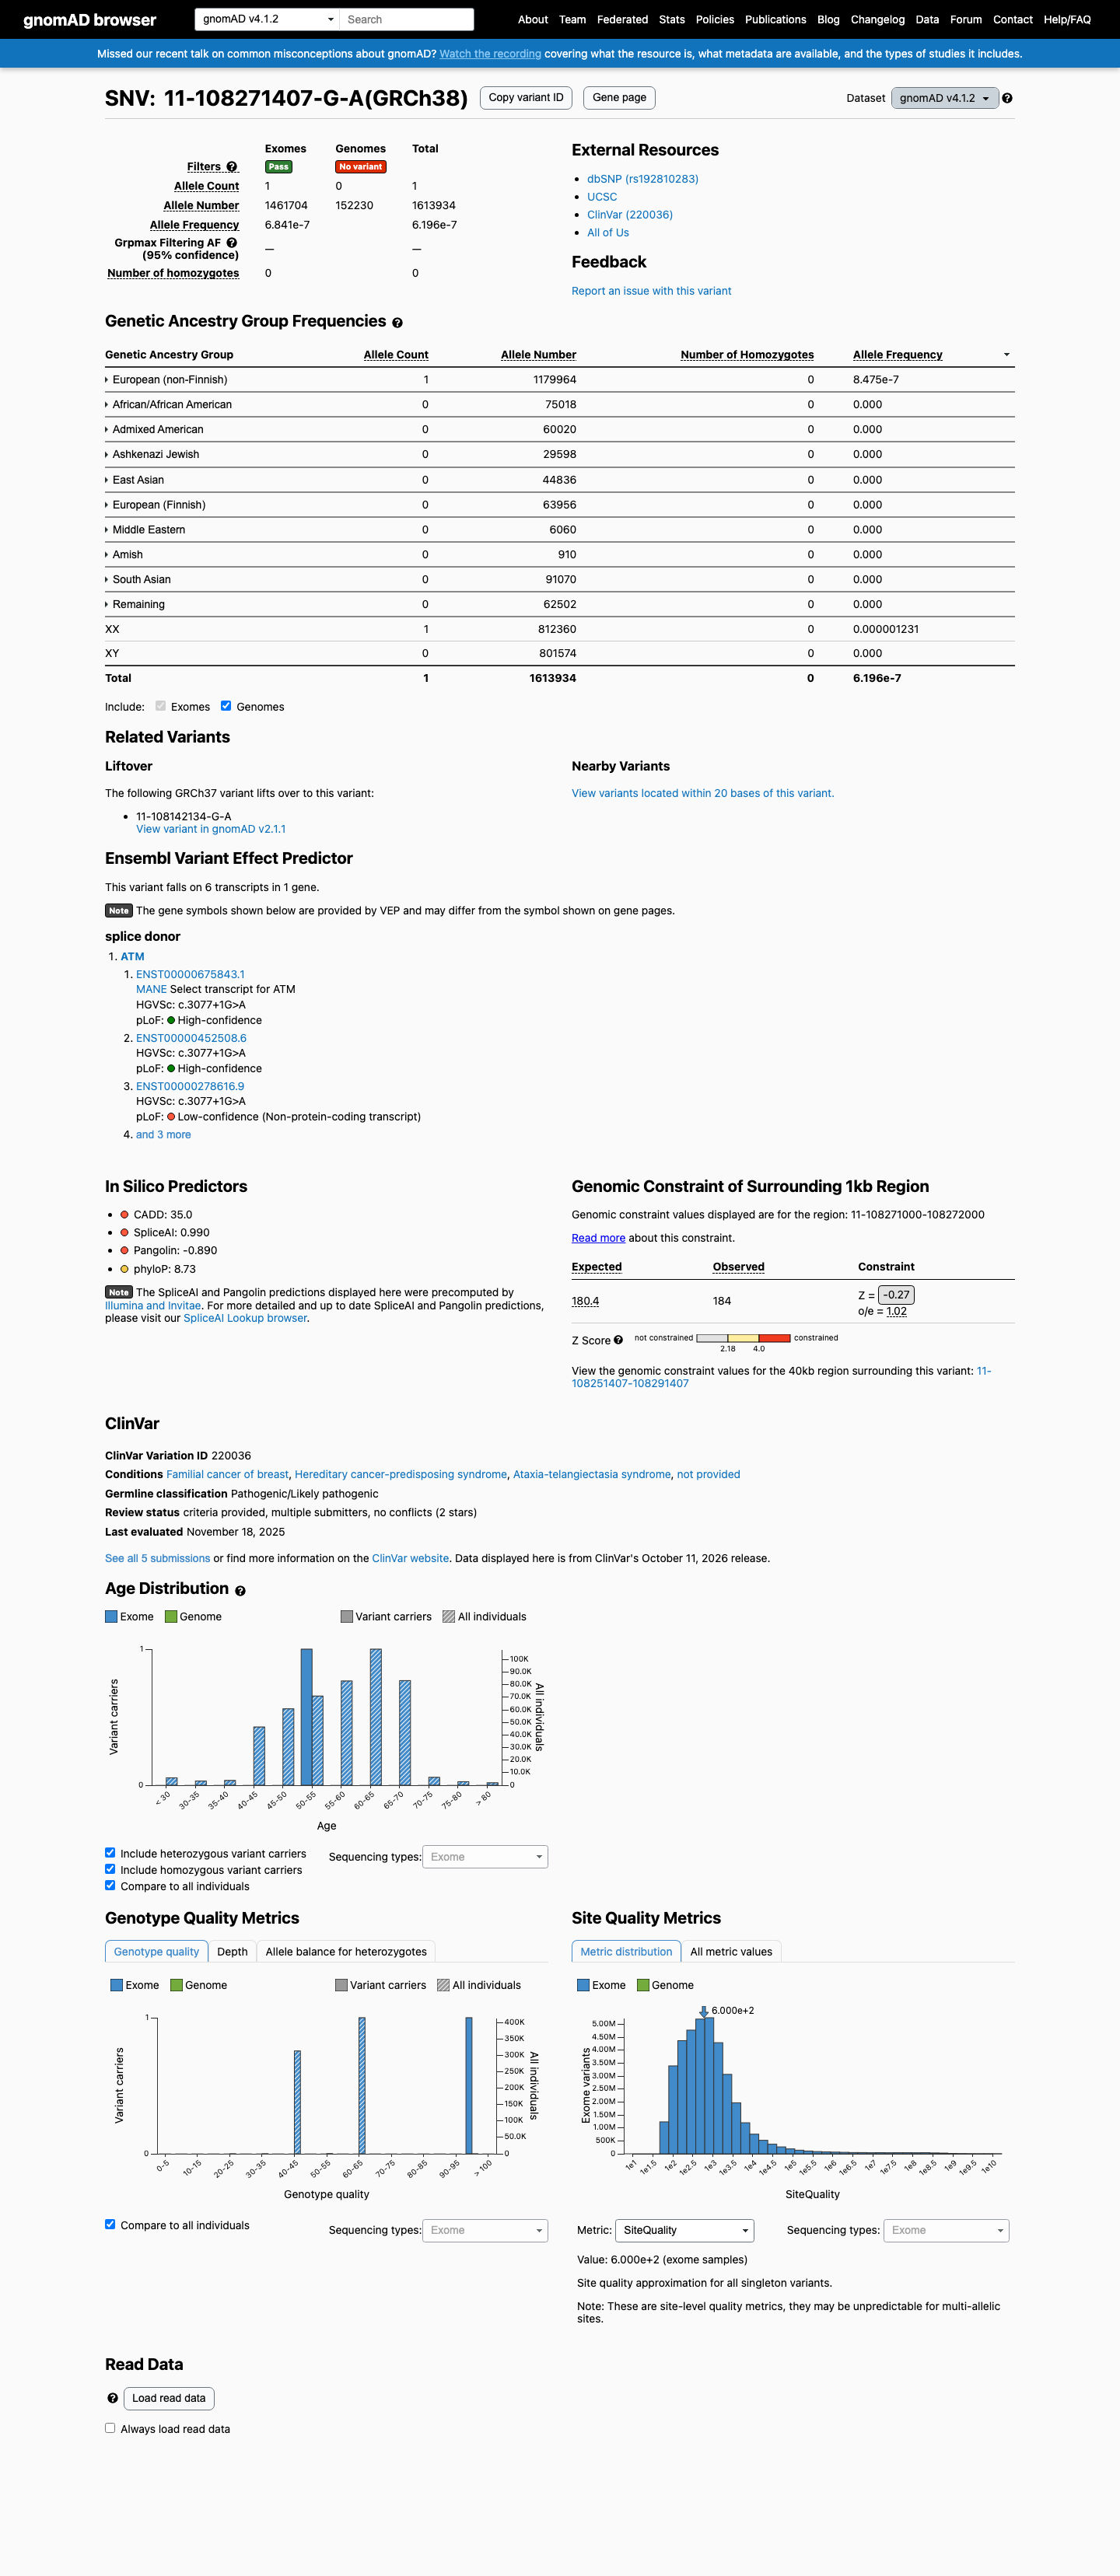
This variant is present in gnomAD v4.1 (AF= 6.19604e-07; MAF= 0.00006%, 1/1613934 alleles, homozygotes = 0) and has highest observed frequency in the European (non-Finnish) population (AF= 8.47483e-07; MAF= 0.00008%, 1/1179964 alleles, homozygotes = 0).
v2.1
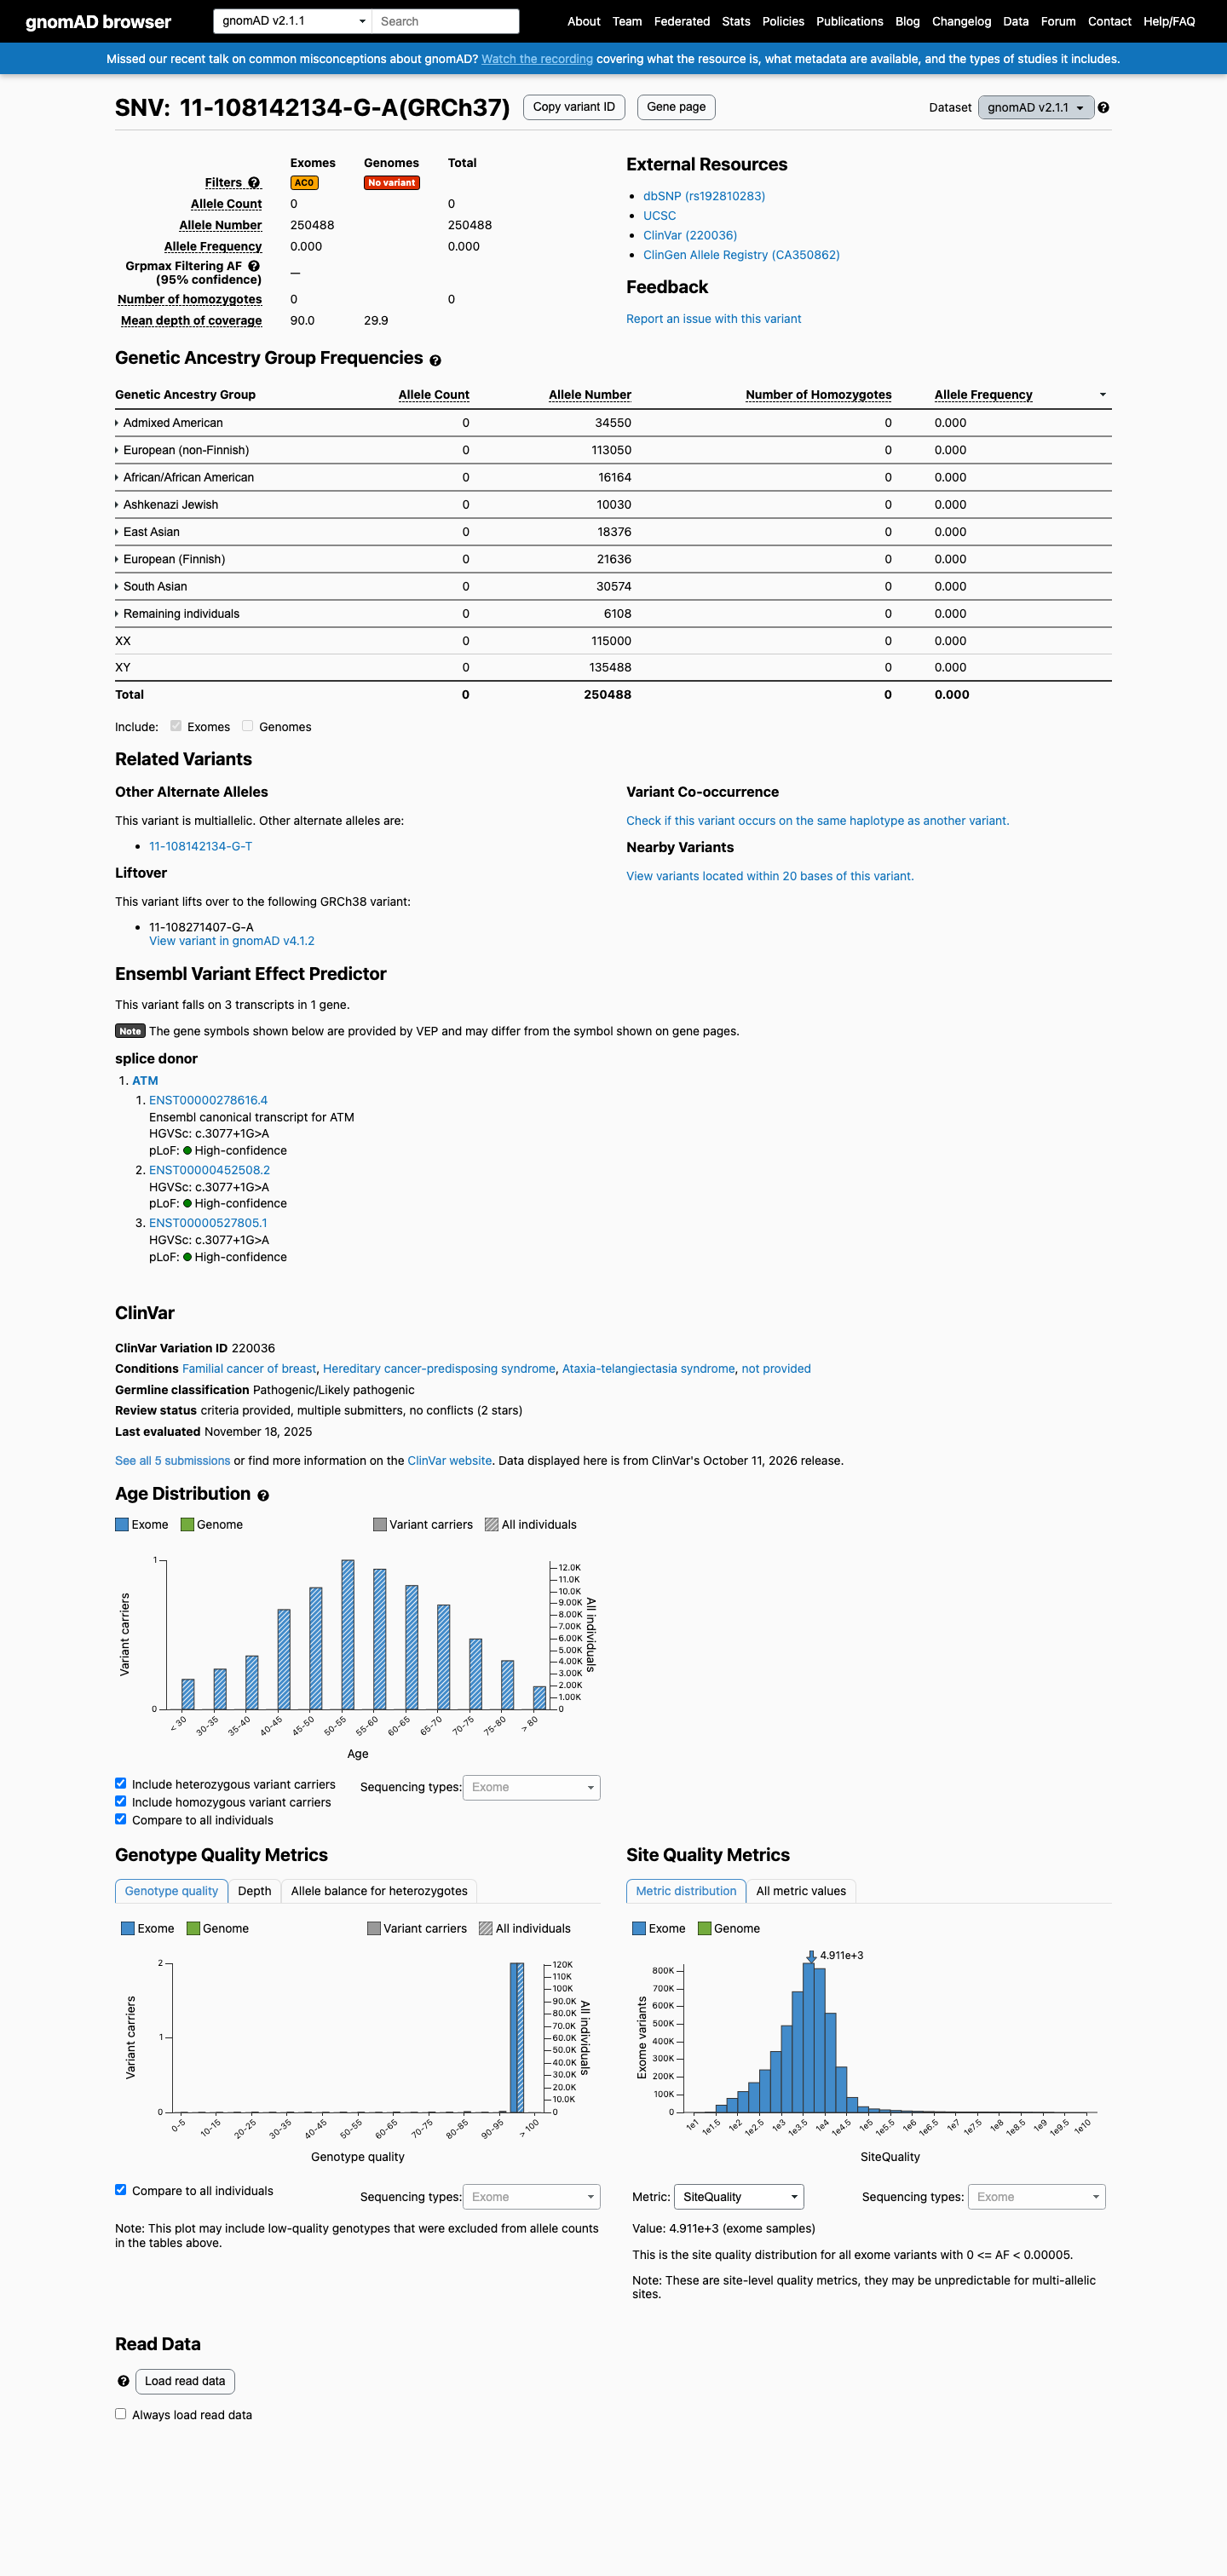
This variant is present in gnomAD v2.1 (AF= 0; MAF= 0.00000%, 0/250488 alleles, homozygotes = 0) and has highest observed frequency in the African/African American population (AF= 0; MAF= 0.00000%, 0/16164 alleles, homozygotes = 0).
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
6.2e-05%
· 1 / 1,613,934
0 hom
European (non-Finnish)
1 / 1,179,964
8.5e-05%
+ 9 not observed (Remaining individuals, Admixed American, European (Finnish), Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American)
gnomAD v2.1
Absent
· 0 / 250,488
0 hom
Not observed in any ancestry group.
+ 8 not observed (African/African American, Admixed American, Ashkenazi Jewish, East Asian, European (Finnish), European (non-Finnish), Remaining individuals, South Asian)
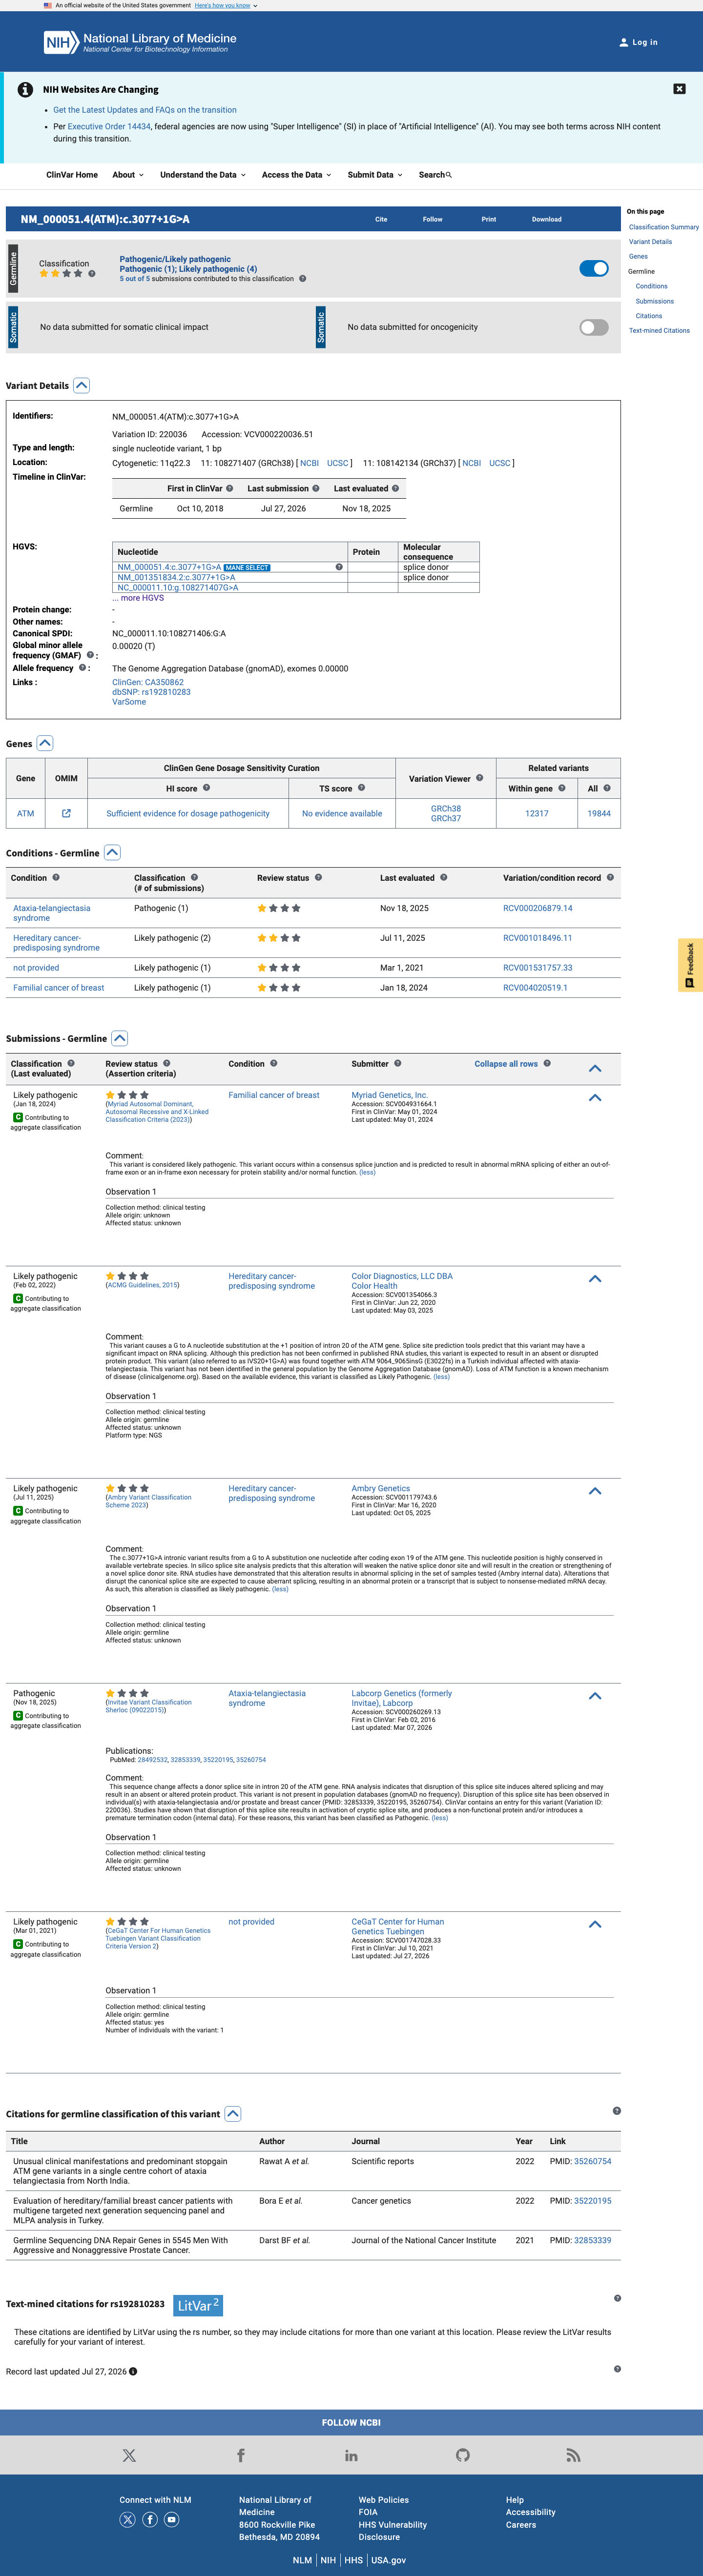
This variant has been reported in ClinVar as Likely pathogenic (4 clinical laboratories) and as Pathogenic (1 clinical laboratory). (ClinVarID = 220036)
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 8 further PMIDs triaged but not cited — see Sources & references.
Sun et al. Supplementary Table S1 contains the exact variant c.3077+1G>A in exon 20 and classifies it as a high-confidence non-functional splice variant from direct prime-editing assessment, with DeepATM_predicted=No.
Variant
✓ Names this variant — characterised directly
Applied to
→PVS1
strong
The exact variant's direct non-functional splice-assay result corroborates the predicted loss-of-function consequence used alongside the VCEP PVS1 rule.
→PS3
supporting
The exact-row direct prime-editing result is Non-functional with High confidence and supports PS3_Supporting under the VCEP file-specific rule.
108271407 G A c.3077+1G>A exon_20 11 Splice_AD 35 NA NA NA NA 6.84133039247344e-07 Likely pathogenic -3.24742909184742 Non-functional High No
Location Supplementary Table S1, converted full-text row 9790 · Context Sun et al. 2025 functional assessment of ATM single-nucleotide variants using prime editing and deep learning; the exact row is marked Non-functional, High confidence, and DeepATM_predicted=No. · full text
Sources & reference links
8Sources
CSpec VCEP
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
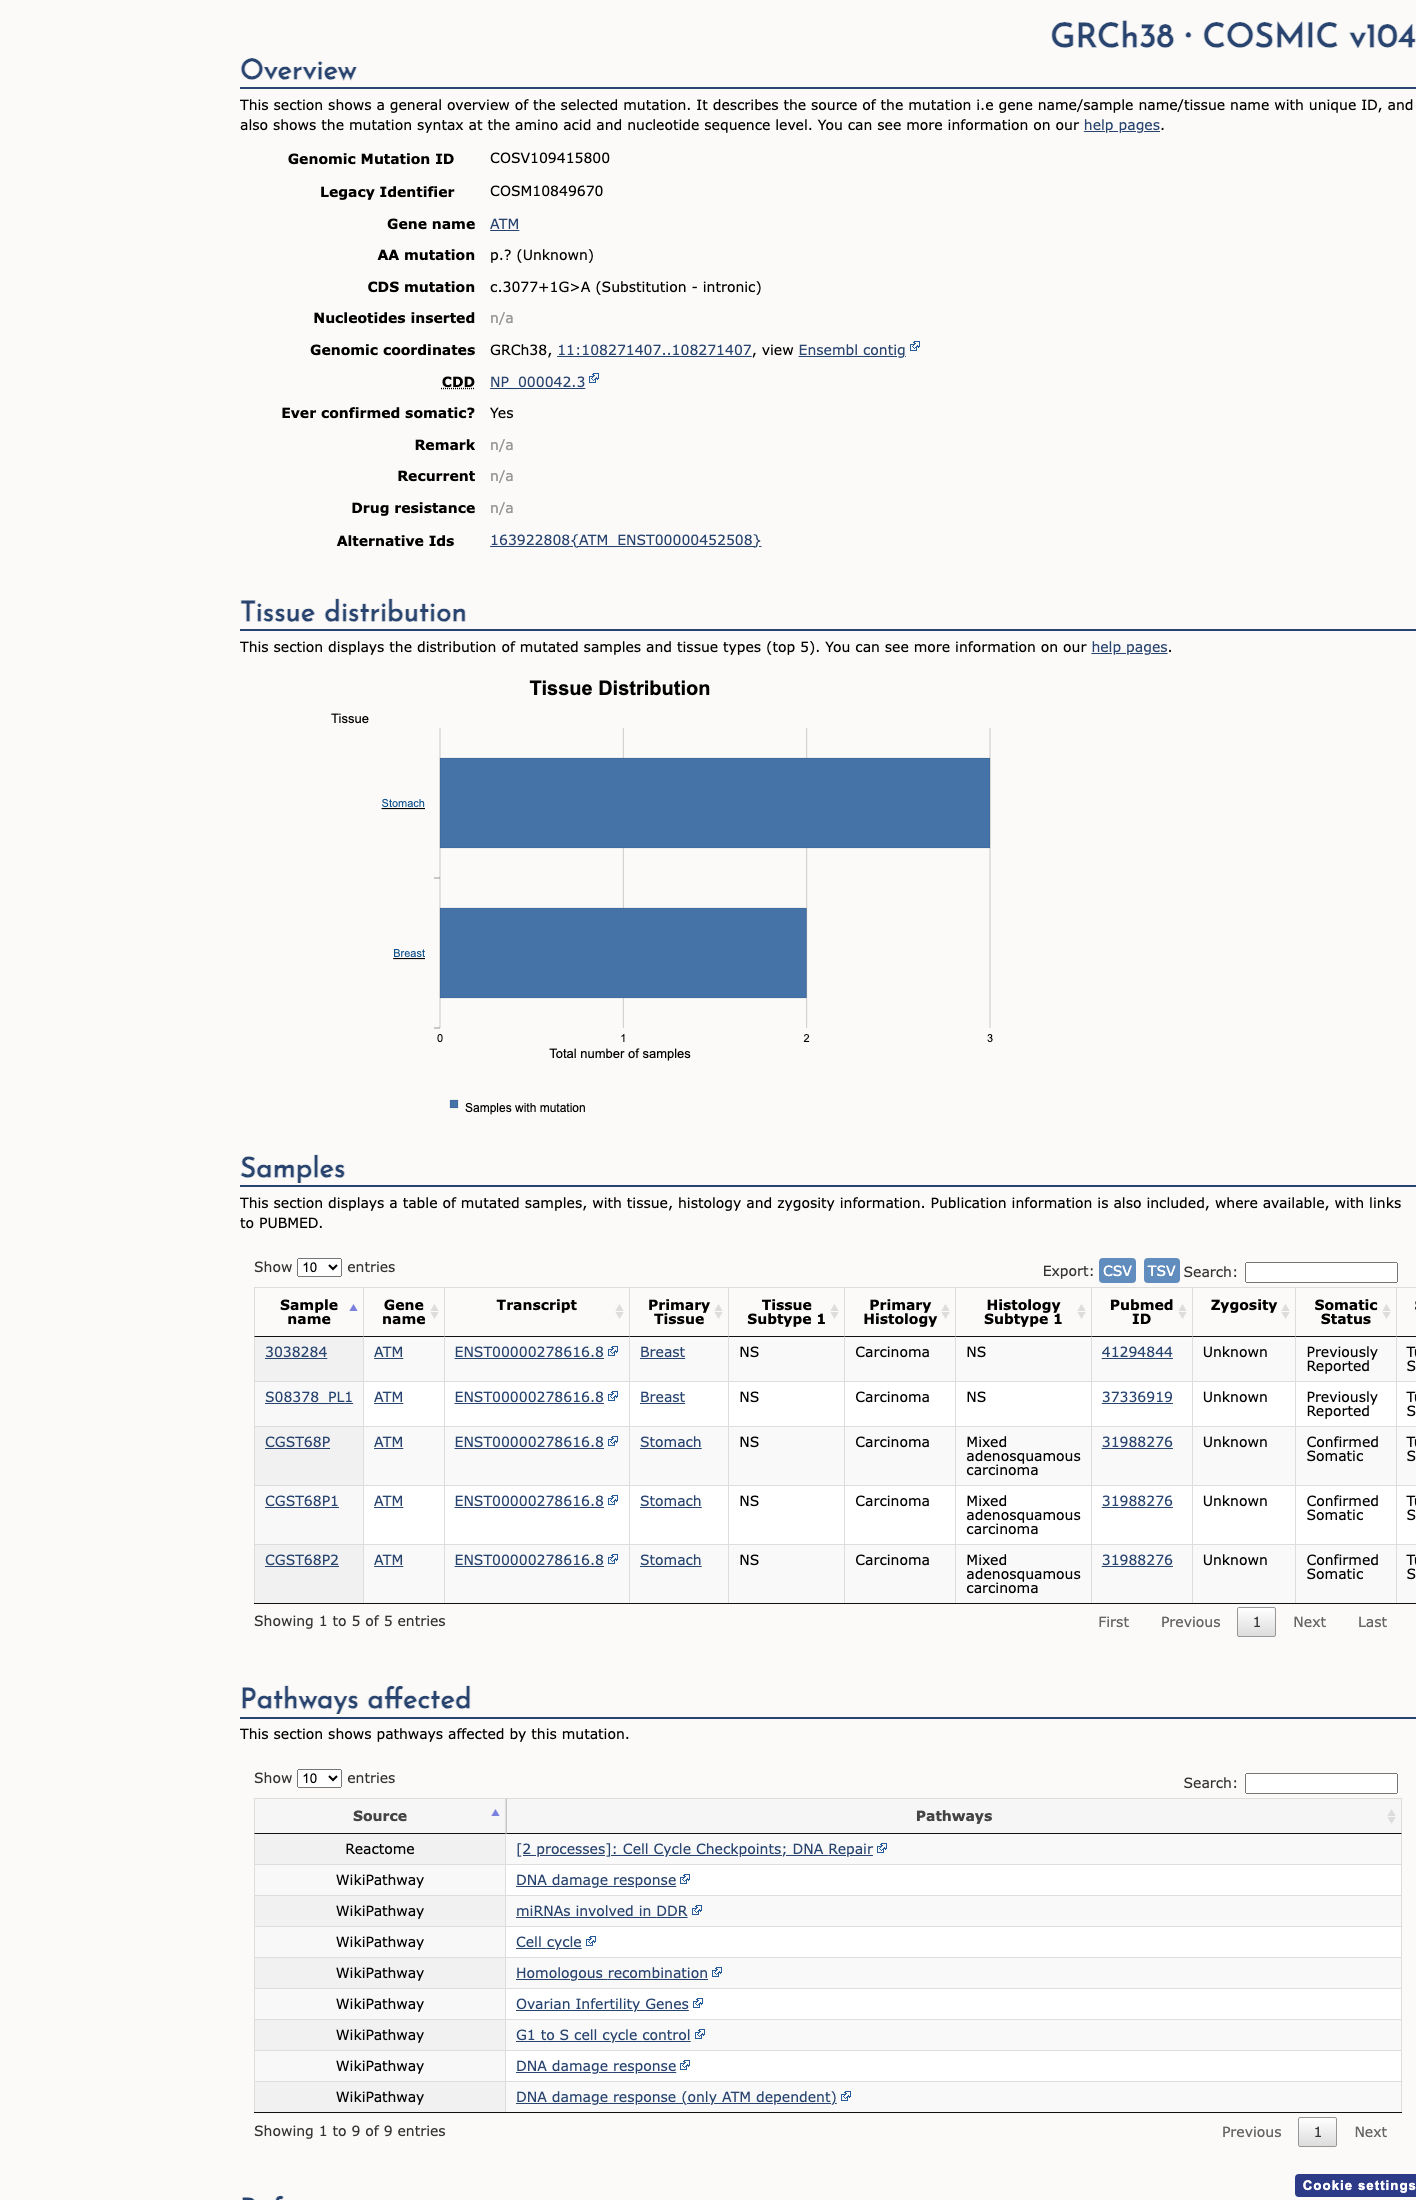
COSMIC
Triaged references · 8 PMIDs not cited in assessment
25741868 ↗Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.CLINVAR
32853339 ↗Germline Sequencing DNA Repair Genes in 5545 Men With Aggressive and Nonaggressive Prostate Cancer.CLINVAR
24366402 ↗Summaries for patients. Assessing the genetic risk for BRCA-related breast or ovarian cancer in women: recommendations from the U.S. Preventive Services Task Force.CLINVAR
24418350 ↗EFNS/ENS Consensus on the diagnosis and management of chronic ataxias in adulthood.CLINVAR
35220195 ↗Evaluation of hereditary/familial breast cancer patients with multigene targeted next generation sequencing panel and MLPA analysis in Turkey.CLINVAR
35260754 ↗Unusual clinical manifestations and predominant stopgain ATM gene variants in a single centre cohort of ataxia telangiectasia from North India.CLINVAR