This variant was interpreted by pipeline
7.1.0.
The current version is
8.0.0.
Re-running queues a fresh interpretation; the result replaces this page when complete.
Re-runs are not free. Blank = draws on today's public interpretation allowance.
With a code = uses 1 of that code's uses.
ATM encodes a kinase that acts as a master controller of the cellular DNA damage response: it detects double-strand breaks and coordinates DNA repair, cell cycle arrest, or apoptosis through downstream targets including p53, BRCA1, and CHK2. Loss-of-function mutations in both copies of ATM cause ataxia telangiectasia, an autosomal recessive disorder characterized by neurological problems, immune deficiency, and cancer predisposition. ATM functions as a tumor suppressor: individuals with ataxia telangiectasia are prone to childhood lymphomas, leukemias, and breast cancer, while carriers of a single altered copy have increased risk of breast, pancreatic, prostate, and other cancers. Somatic ATM mutations also occur in lymphoid malignancies and solid tumors, and ATM-deficient cancers are often especially sensitive to DNA-damaging treatments.
This variant
ATM is a tumor suppressor whose loss of function causes ataxia telangiectasia and raises cancer risk in carriers. This missense change is classified as a variant of uncertain significance: it is extremely rare in the general population, but no functional, segregation, or clinical evidence currently shows it impairs ATM's DNA-damage-response function or predisposes to disease.
Transcript
NM_000051.4
HGVS · transcript:coding
NM_000051.4:c.3332T>C
GRCh38
chr11:108279538 T>C
GRCh37
chr11:108150265 T>C
Only PM2 (supporting) was met under the ATM VCEP v1.5 criteria-combination framework - insufficient evidence for any pathogenic or benign classification.
Classification rationale
PM2VUS
ATM c.3332T>Cmissense · exon 23
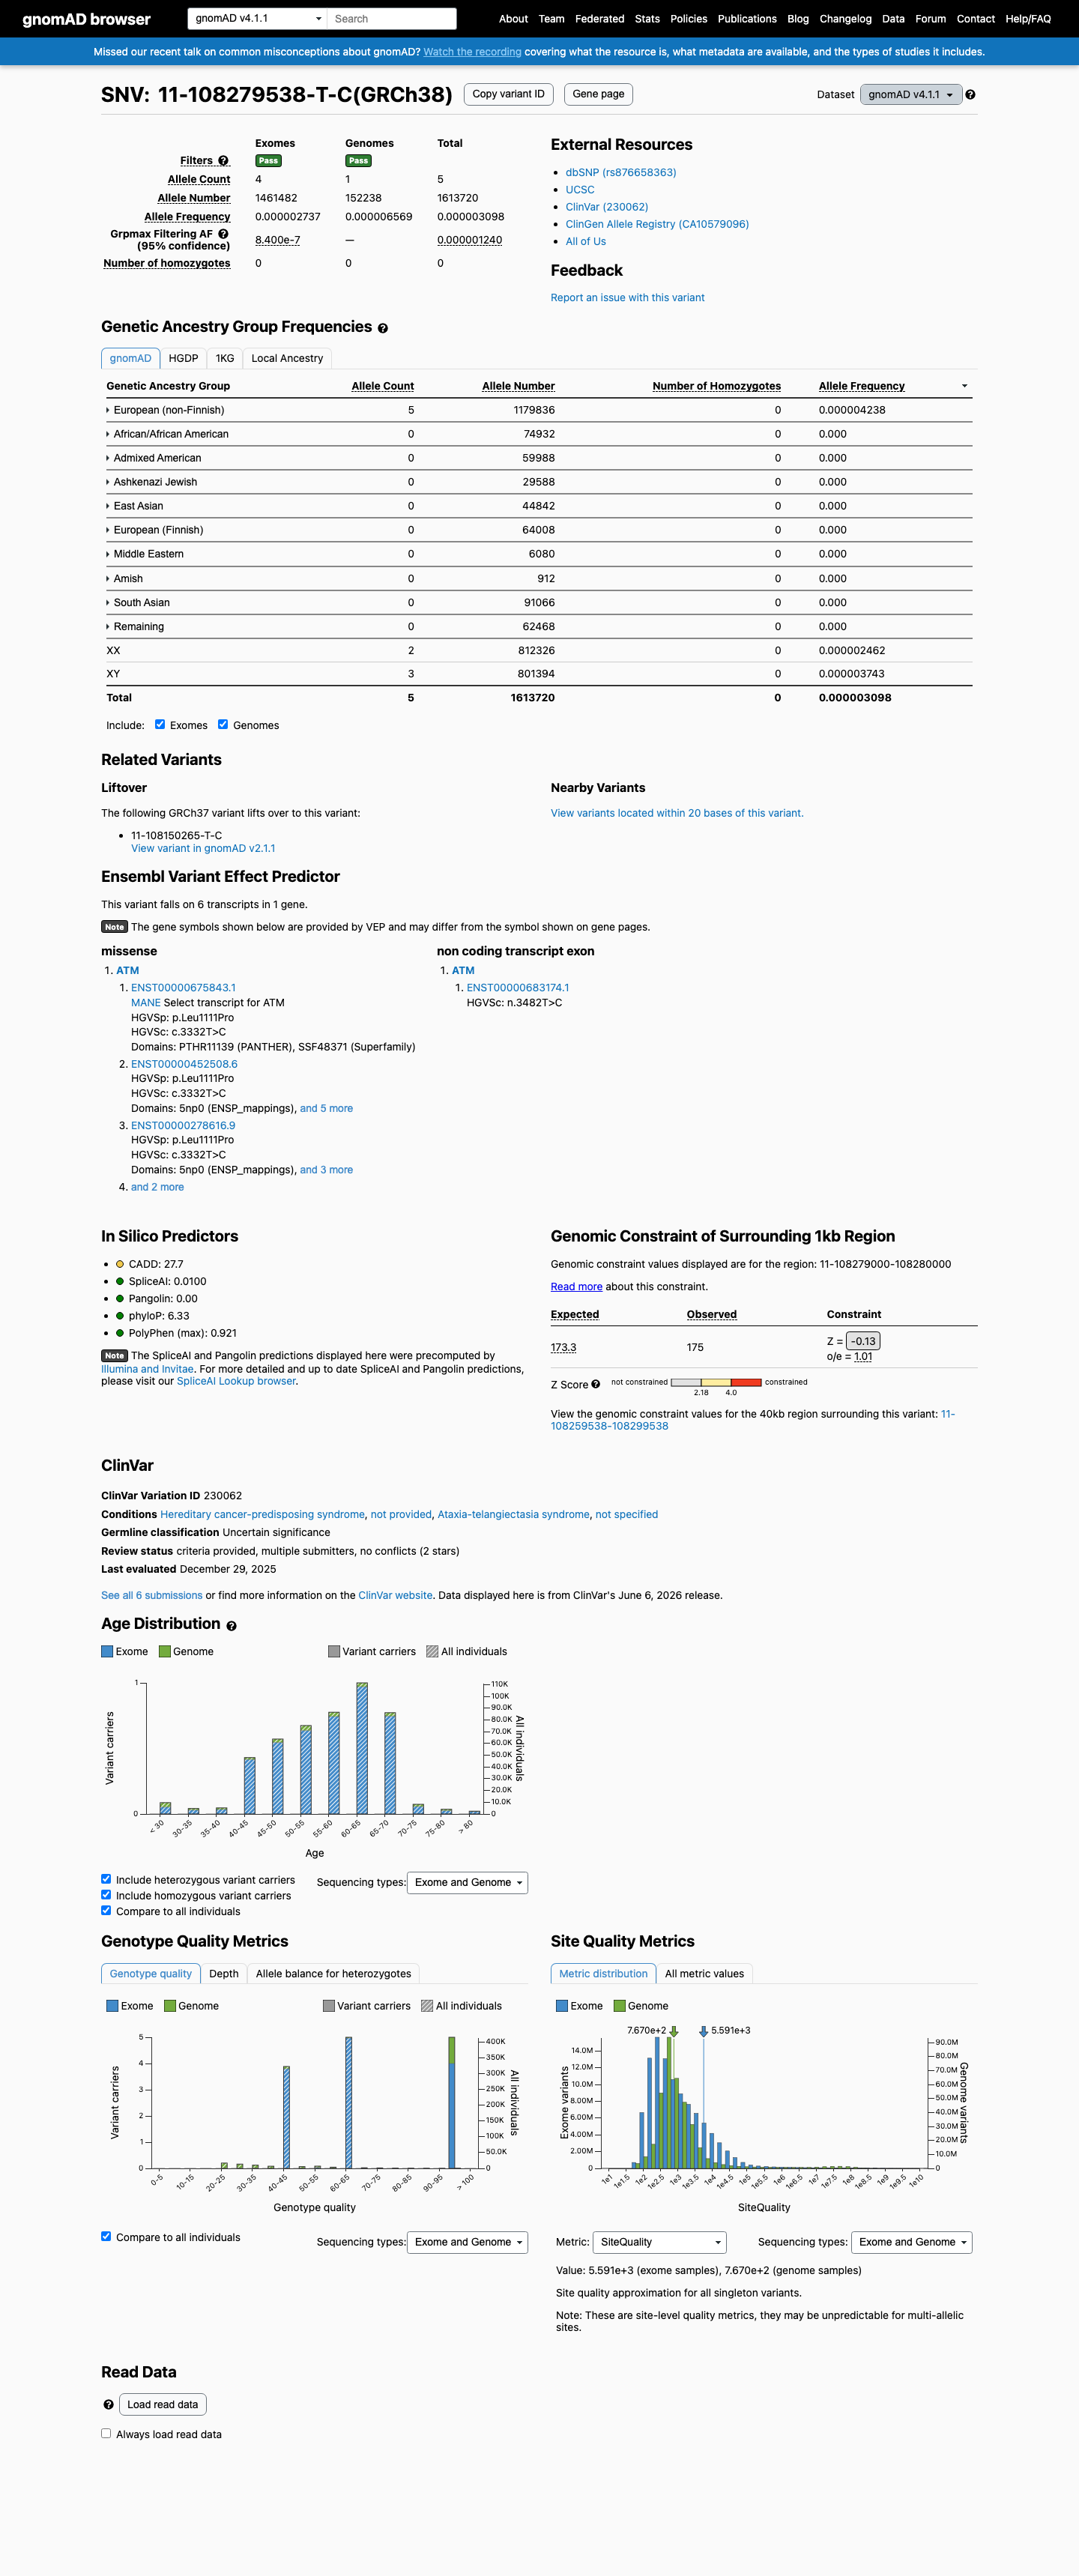
PM2 (Supporting): gnomAD v4.1 allele frequency 0.00031% (3.1e-06), below the <=0.001% threshold, with no observed homozygotes. Overall classification: VUS - only PM2 (Supporting) was met, insufficient for any pathogenic or benign rule under the ATM VCEP v1.5 combination framework.
PM2→VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria
may be applied incorrectly, sources may be misread, and a confident-looking
classification can still be wrong. Double-check every criterion and
its underlying evidence before relying on any call.
Gene diagram
· NM_000051.4 · variants mapped to exon structure
ATMNM_000051.4
Fetching transcript structure from UCSC…
Exons
—
Transcript span
—
Strand
—
Variants mapped
—
Source
UCSC ncbiRefSeqCurated
All variants in ATM—click a row to locate it on the plot · use the link column to open its page
Protein
Location
Classification
Link
Applied criteria · 1 applied · 11 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 1
Strength Supporting Moderate Strong Very strong
✓
PM2supportingPathogenic
Met (supporting): gnomAD v4.1 allele frequency 0.00031% (3.1e-06), below the <=0.001% PM2 threshold, with no observed homozygotes.
ClinGen HBOP ATM VCEP v1.5 specifies PM2_Supporting at frequency <=0.001% in gnomAD v4.1 and states that n=1 in a single subpopulation is sufficiently rare.gnomAD v4.1 reports 5/1,613,720 alleles overall (AF 3.09843e-06; 0.000309843%), a highest population AF of 4.237877e-06 in European non-Finnish individuals (5/1,179,836), grpmax FAF 1.24e-06, and zero homozygotes.The variant is absent from gnomAD-Canada v1.0, providing additional population rarity context without changing the VCEP threshold assessment.
Assessed · not applied
· 5 not met · 6 not assessed
Pathogenic
PS1Not met: no established pathogenic or likely pathogenic variant producing the same p.Leu1111Pro amino-acid change was found.
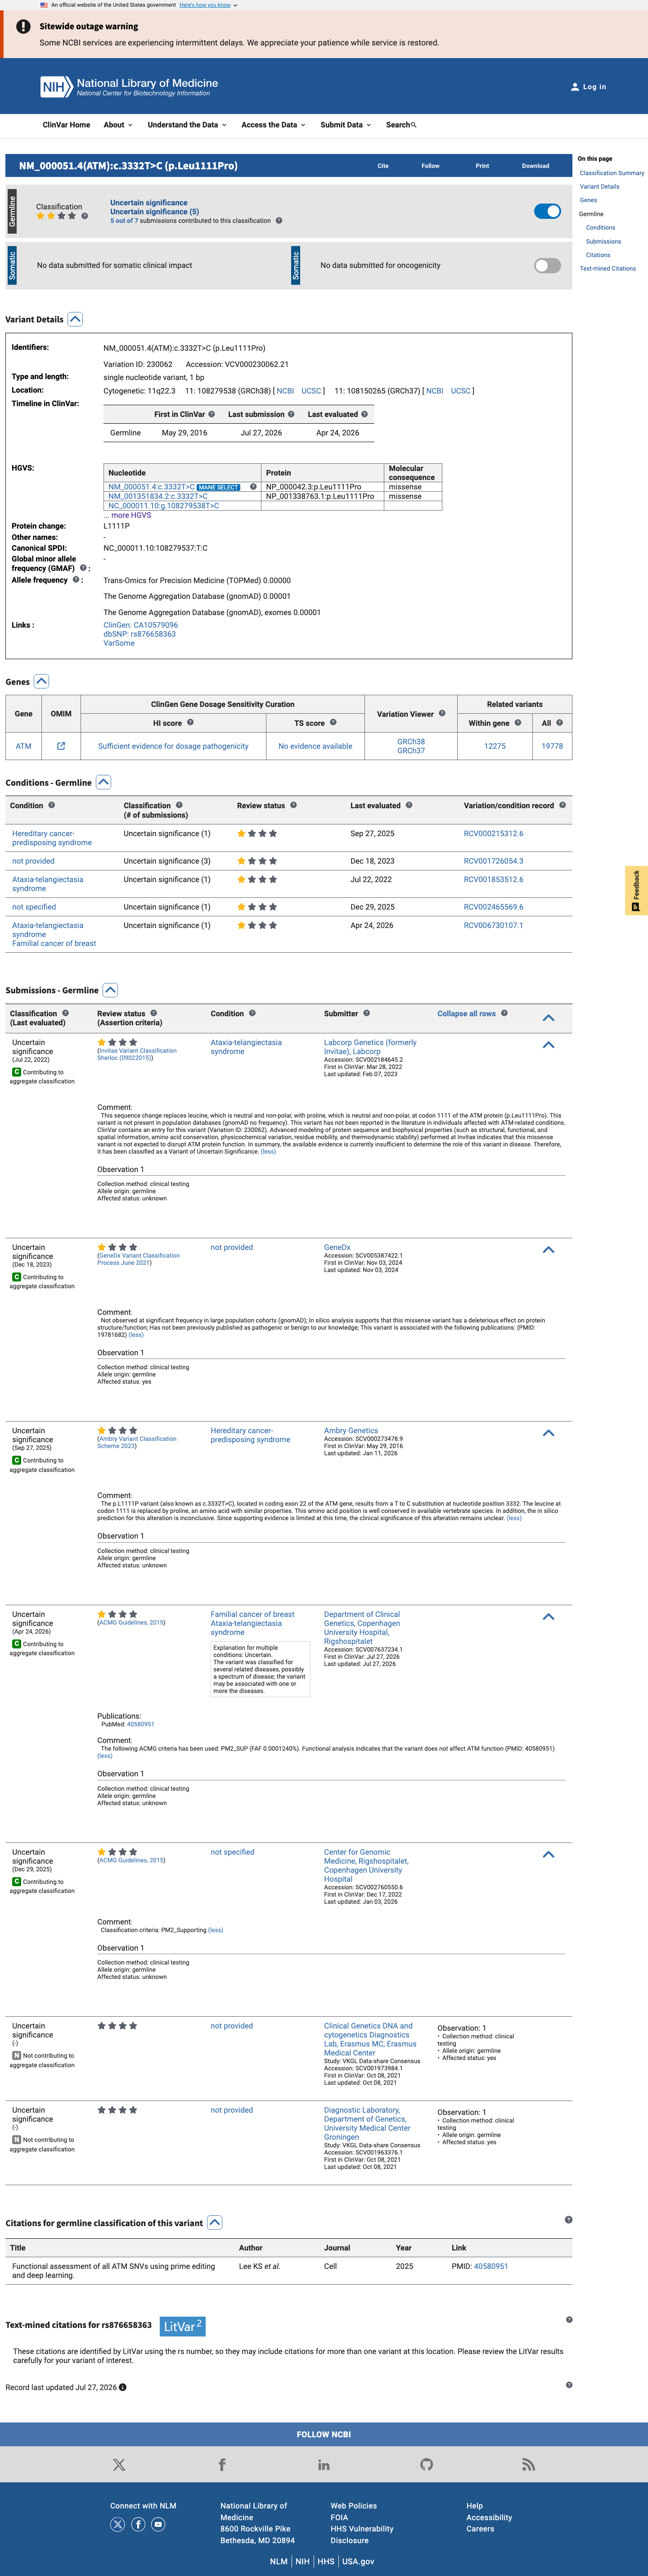
PS3Not assessed: no VCEP-approved assay result exists; a 2025 screen reporting normal function is flagged for human review as it is not VCEP-calibrated.
PS4Not assessed: no case-control study with a qualifying odds ratio or p-value for this variant was available.
PM3Not assessed: no affected-proband observation or second pathogenic ATM variant in trans was documented.
PP1Not assessed: no affected relatives, parental testing, or other segregation observations were documented.
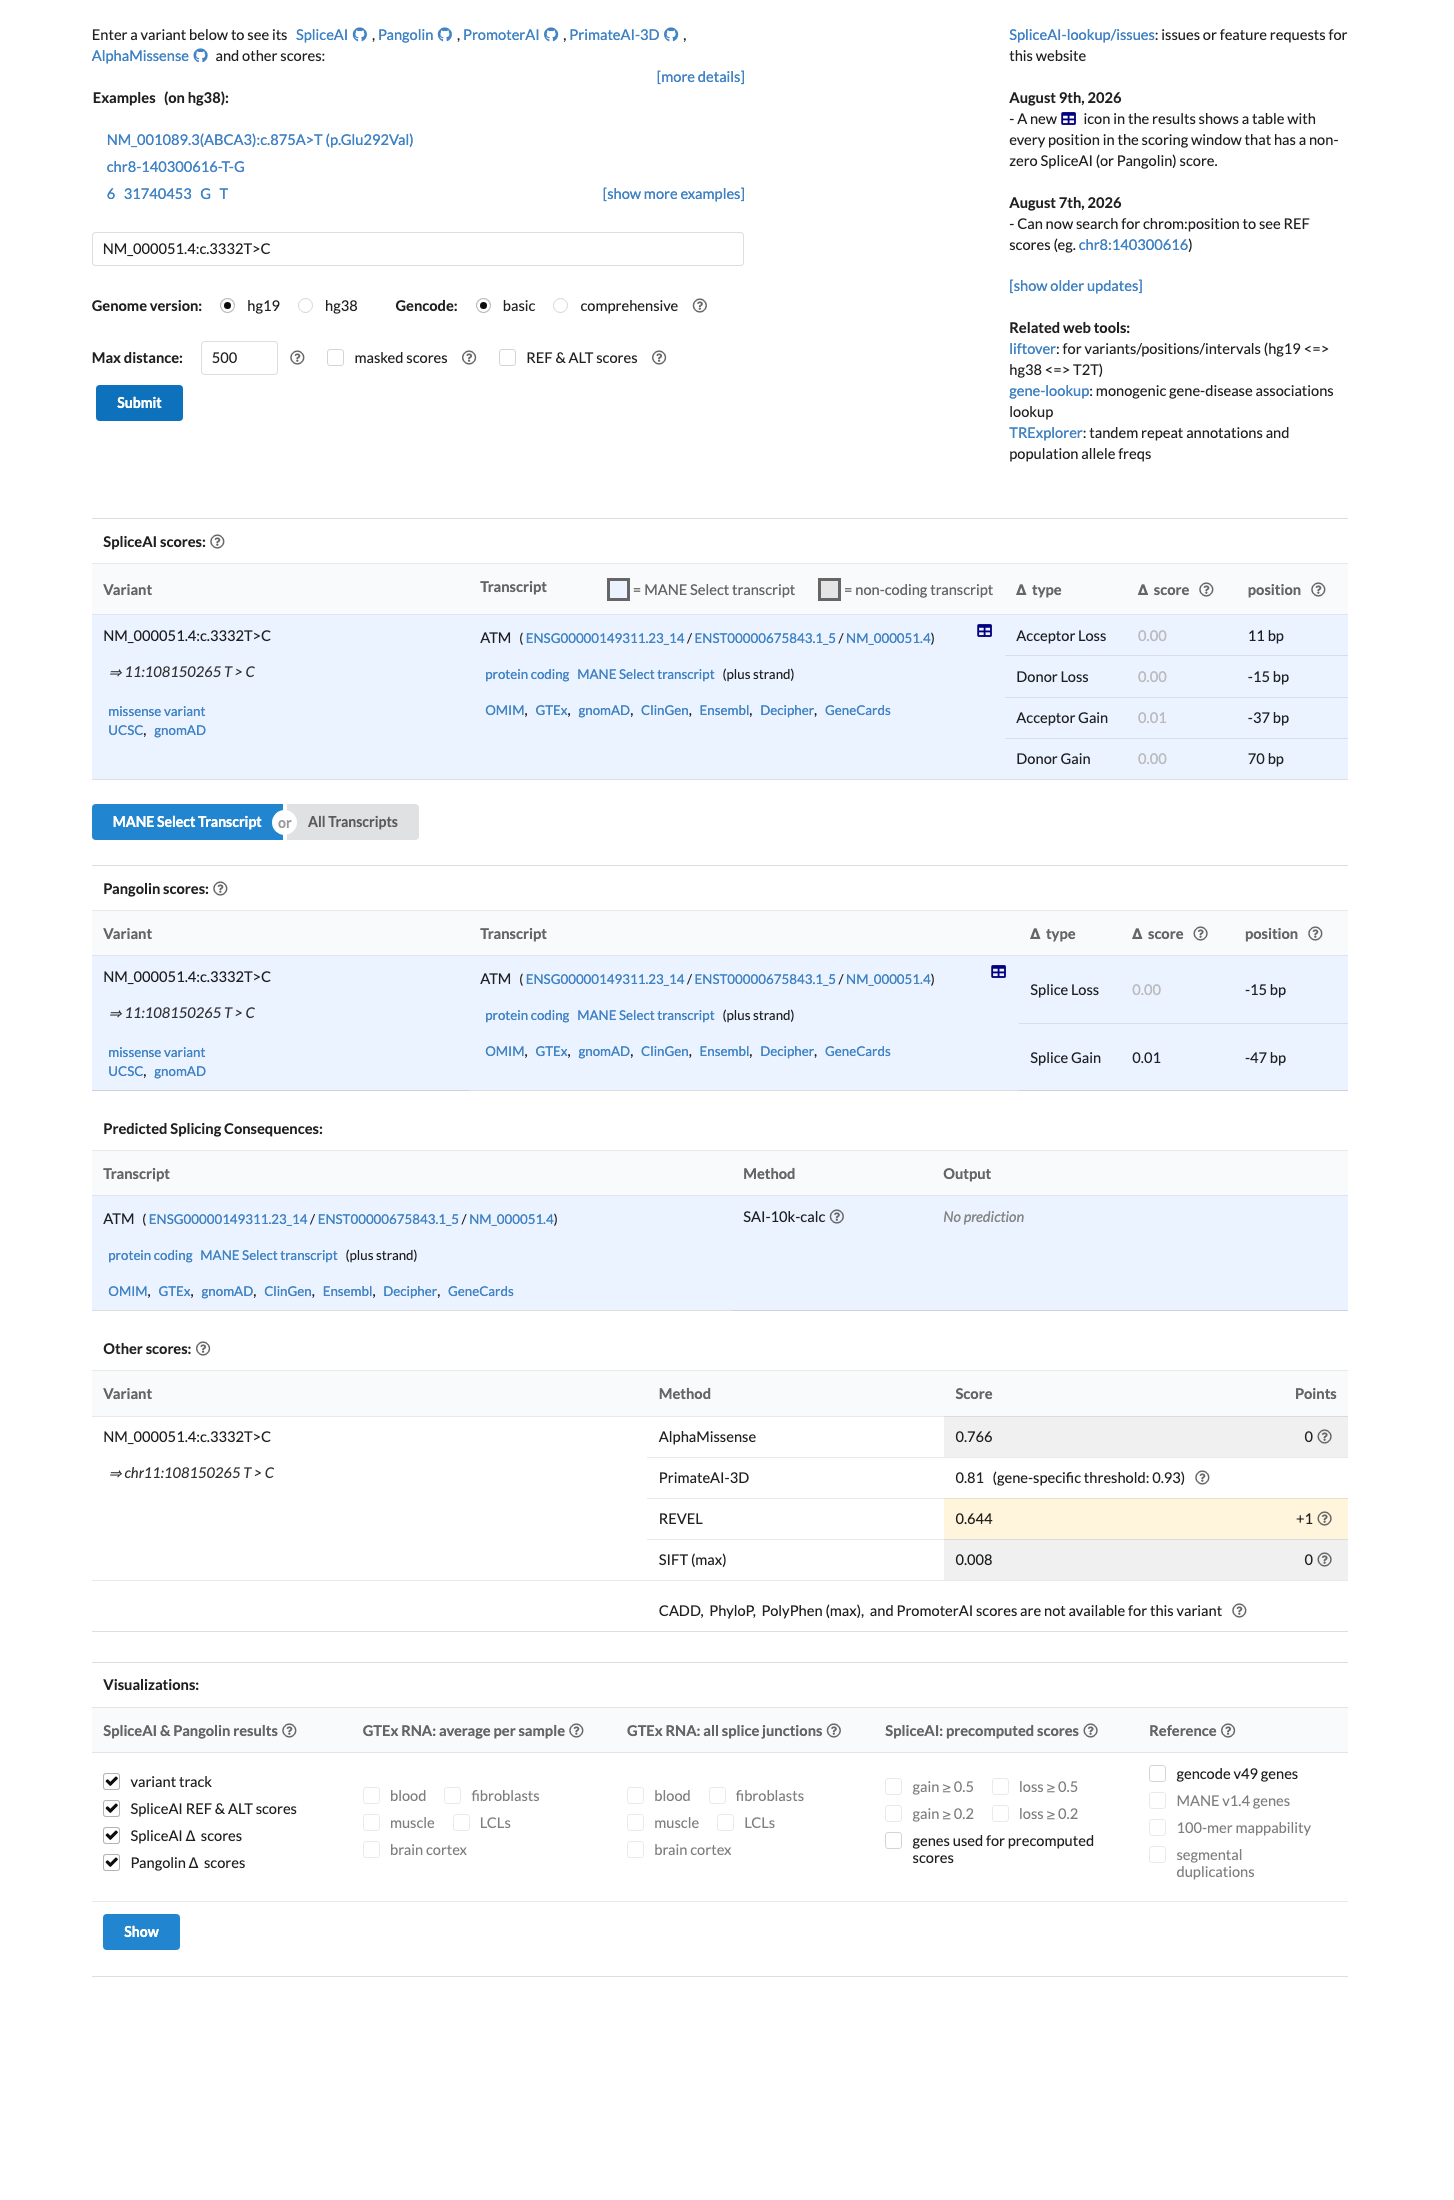
PP3Not met: REVEL 0.644, below the >0.7333 threshold required for missense variants.
Benign
BA1Not met: grpmax filtering allele frequency 0.000124% (1.24e-06), far below the >0.5% BA1 threshold.
BS1Not met: grpmax filtering allele frequency 0.000124% (1.24e-06), below the >0.05% BS1 threshold.
BS3Not assessed: no VCEP-approved assay result exists; a 2025 screen reporting normal function is flagged for human review as it is not VCEP-calibrated.
BP2Not assessed: no unaffected carrier with a pathogenic ATM variant in trans or phase information was documented.
BP4Not met: REVEL 0.644 lies above the <=0.249 BP4 threshold (gray zone between BP4 and PP3).
This variant is present in gnomAD v4.1 (AF= 3.09843e-06; MAF= 0.00031%, 5/1613720 alleles, homozygotes = 0) and has highest observed frequency in the European (non-Finnish) population (AF= 4.23788e-06; MAF= 0.00042%, 5/1179836 alleles, homozygotes = 0); grpmax FAF= 1.24e-06.
v2.1
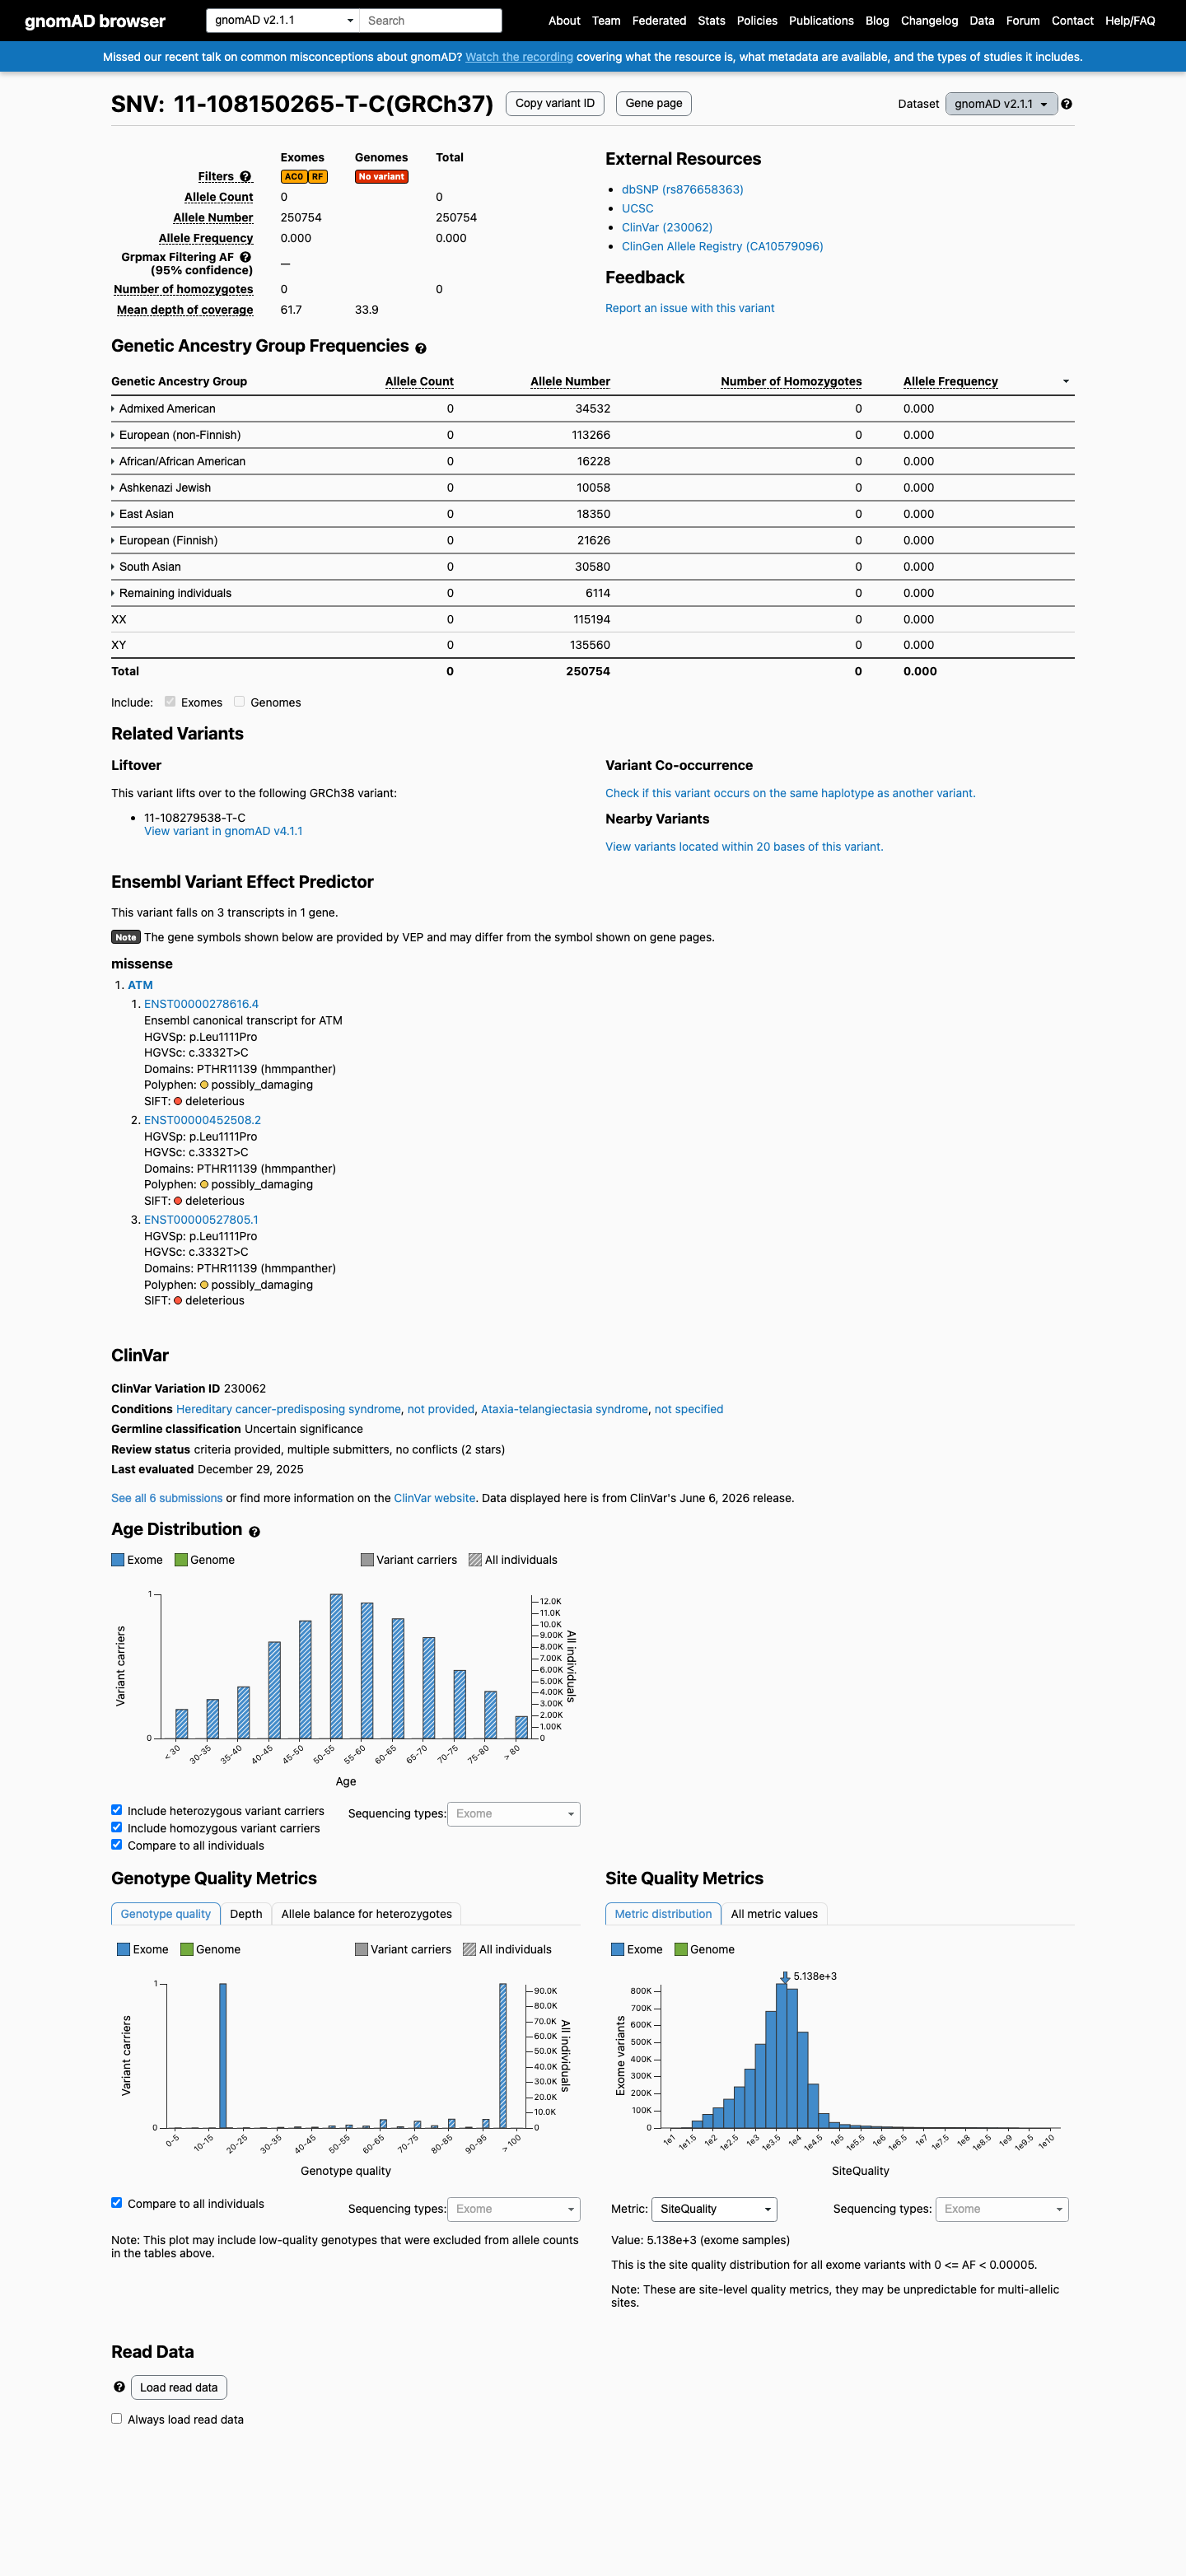
This variant is present in gnomAD v2.1 (AF= 0; MAF= 0.00000%, 0/250754 alleles, homozygotes = 0) and has highest observed frequency in the African/African American population (AF= 0; MAF= 0.00000%, 0/16228 alleles, homozygotes = 0).
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.00031%
· 5 / 1,613,720
0 hom · FAF 0.00012%
European (non-Finnish)
5 / 1,179,836
0.00042%
+ 9 not observed (Remaining individuals, Admixed American, European (Finnish), Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American)
gnomAD v2.1
Absent
· 0 / 250,754
0 hom
Not observed in any ancestry group.
+ 8 not observed (African/African American, Admixed American, Ashkenazi Jewish, East Asian, European (Finnish), European (non-Finnish), Remaining individuals, South Asian)
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. ATM, a kinase involved in the DNA damage response, is mutated in various solid and hematologic malignancies.
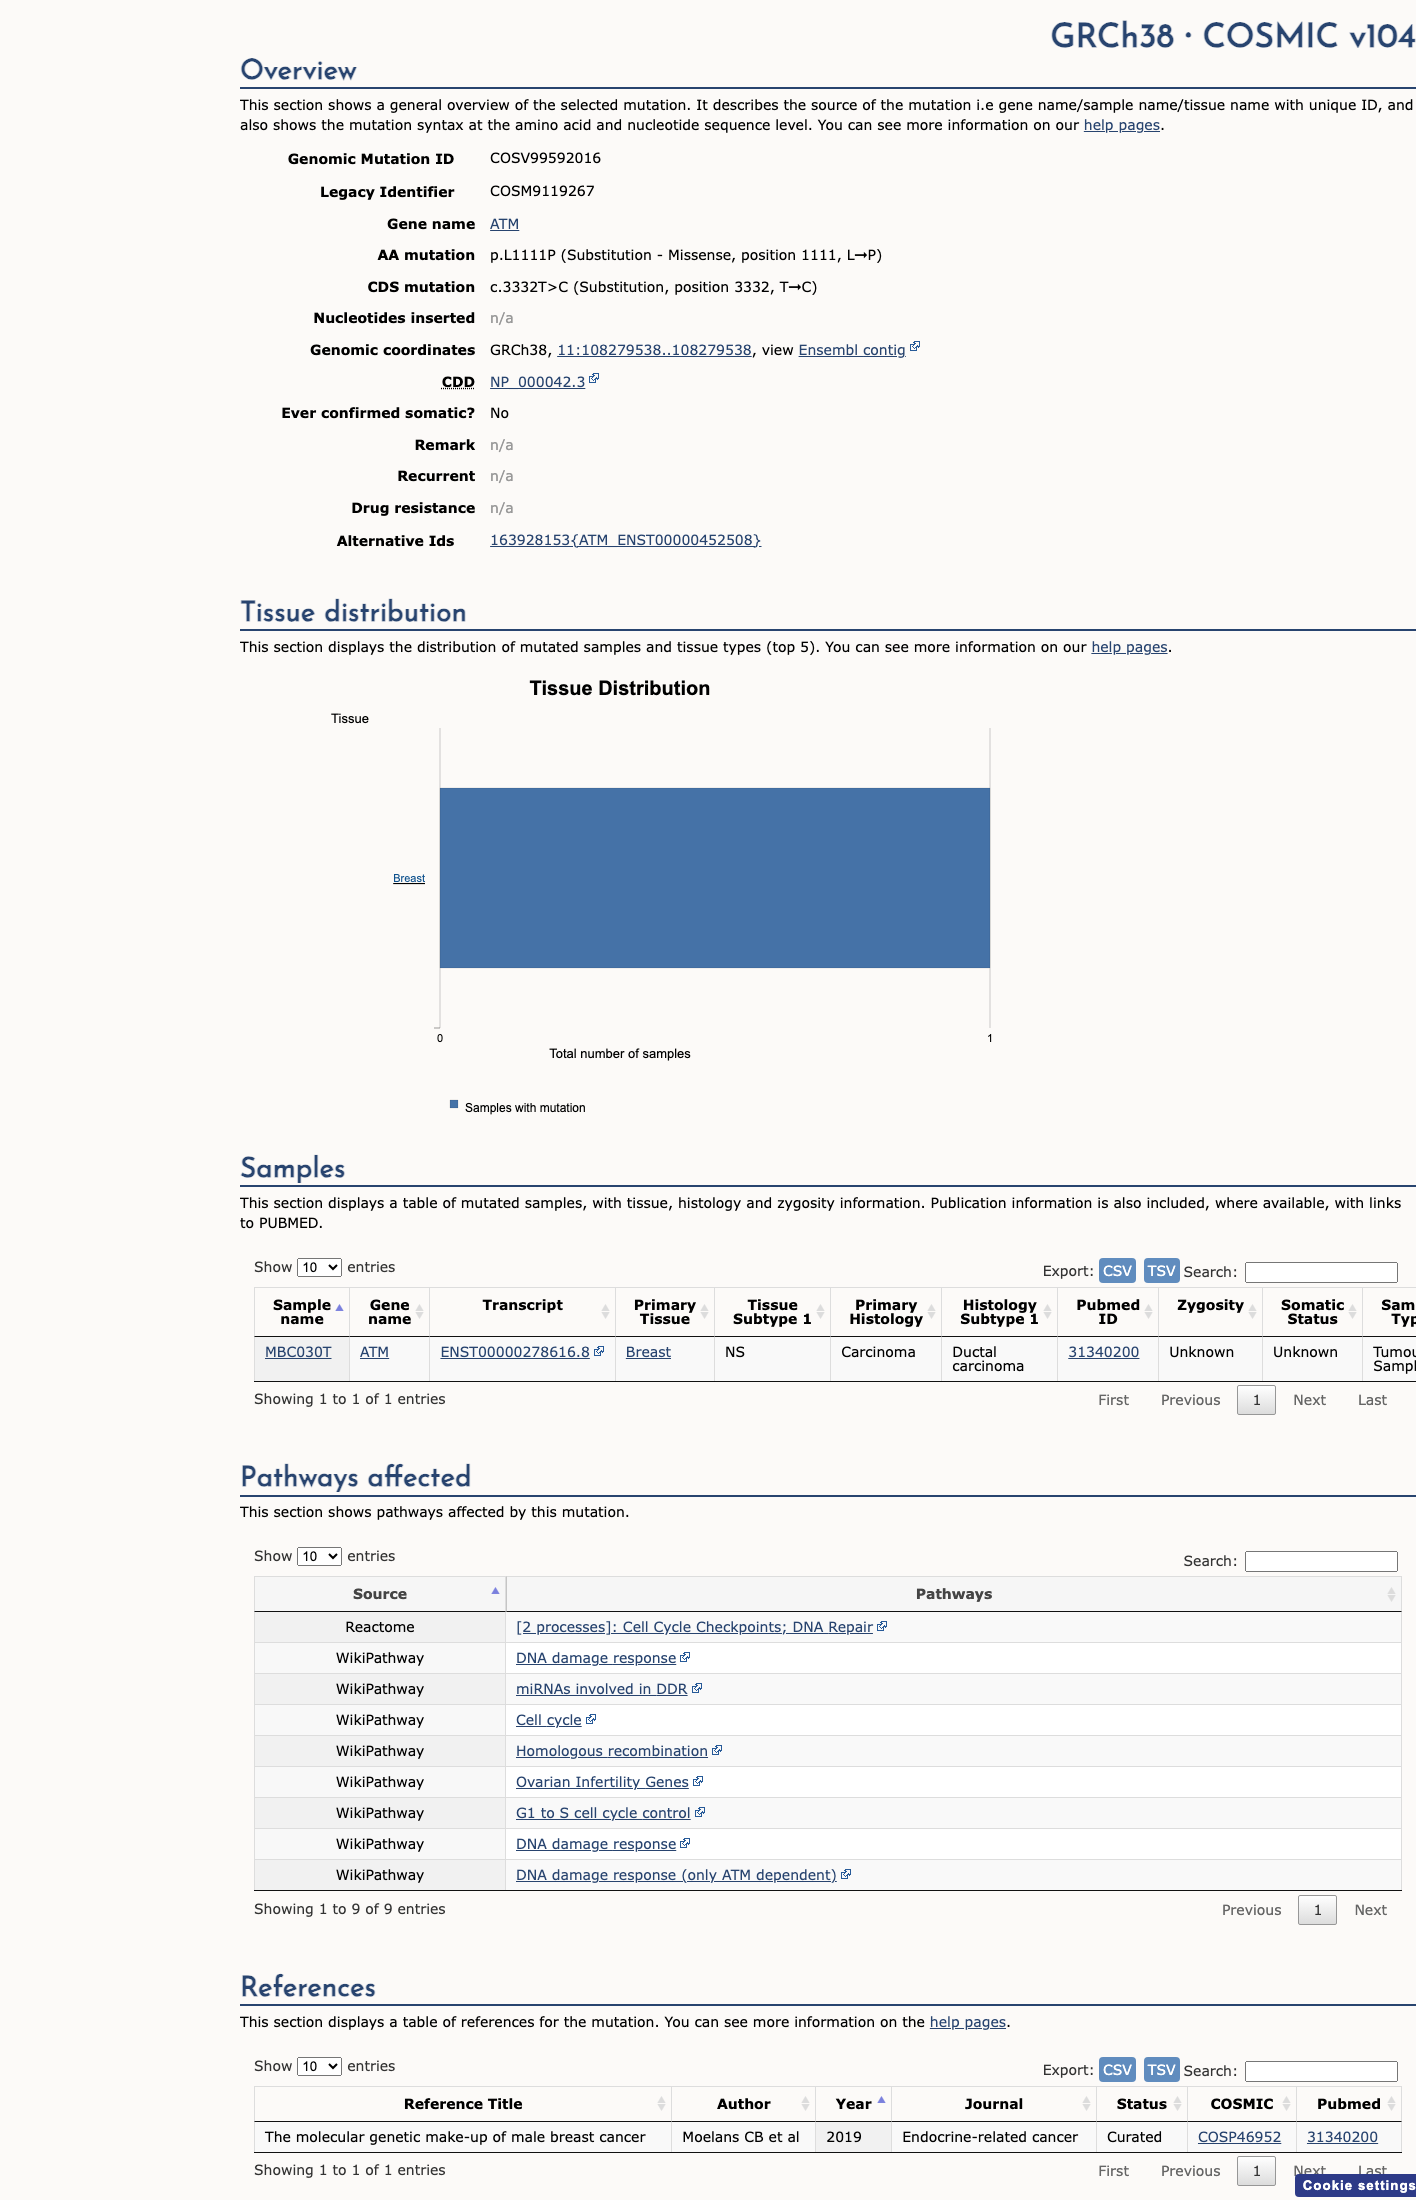
This variant does not lie in a statistically significant hotspot. This variant has previously been reported in somatic cancers (COSMIC; COSV99592016, n = 1 times).
Hotspots
This variant does not lie in a statistically significant hotspot.
Triaged references · 7 PMIDs not cited in assessment
25741868 ↗Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.CLINVAR
40580951 ↗Functional assessment of all ATM SNVs using prime editing and deep learning.CLINVAR
25394175 ↗A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment.CLINVAR
31429903 ↗Risk Assessment, Genetic Counseling, and Genetic Testing for BRCA-Related Cancer: US Preventive Services Task Force Recommendation Statement.CLINVAR
35802134 ↗ACMG SF v3.1 list for reporting of secondary findings in clinical exome and genome sequencing: A policy statement of the American College of Medical Genetics and Genomics (ACMG).CLINVAR