This variant was interpreted by pipeline
7.1.0.
The current version is
8.0.0.
Re-running queues a fresh interpretation; the result replaces this page when complete.
Re-runs are not free. Blank = draws on today's public interpretation allowance.
With a code = uses 1 of that code's uses.
ATM encodes a kinase that acts as a master controller of the cellular DNA damage response: it detects double-strand breaks and coordinates DNA repair, cell cycle arrest, or apoptosis through downstream targets including p53, BRCA1, and CHK2. Loss-of-function mutations in both copies of ATM cause ataxia telangiectasia, an autosomal recessive disorder characterized by neurological problems, immune deficiency, and cancer predisposition. ATM functions as a tumor suppressor: individuals with ataxia telangiectasia are prone to childhood lymphomas, leukemias, and breast cancer, while carriers of a single altered copy have increased risk of breast, pancreatic, prostate, and other cancers. Somatic ATM mutations also occur in lymphoid malignancies and solid tumors, and ATM-deficient cancers are often especially sensitive to DNA-damaging treatments.
This variant
ATM is a tumor suppressor whose loss-of-function mutations cause ataxia telangiectasia and predispose carriers to breast, pancreatic, and other cancers, so variants here are directly relevant to hereditary cancer risk. This p.Ile280Val missense change is a VUS: it is extremely rare in the population yet computationally benign-leaning, so its impact on ATM's DNA-damage-response function remains unresolved and it neither confirms nor excludes disease risk.
Transcript
NM_000051.4
HGVS · transcript:coding
NM_000051.4:c.838A>G
GRCh38
chr11:108244963 A>G
GRCh37
chr11:108115690 A>G
Under the ClinGen HBOP ATM VCEP v1.5 framework exactly two criteria were met - PM2 supporting (rarity) and BP4 supporting (benign-leaning) - and their conflicting directions trigger the conflicting-evidence rule, yielding VUS (uncertain significance).
Classification rationale
PM2BP4VUS
ATM c.838A>Gmissense
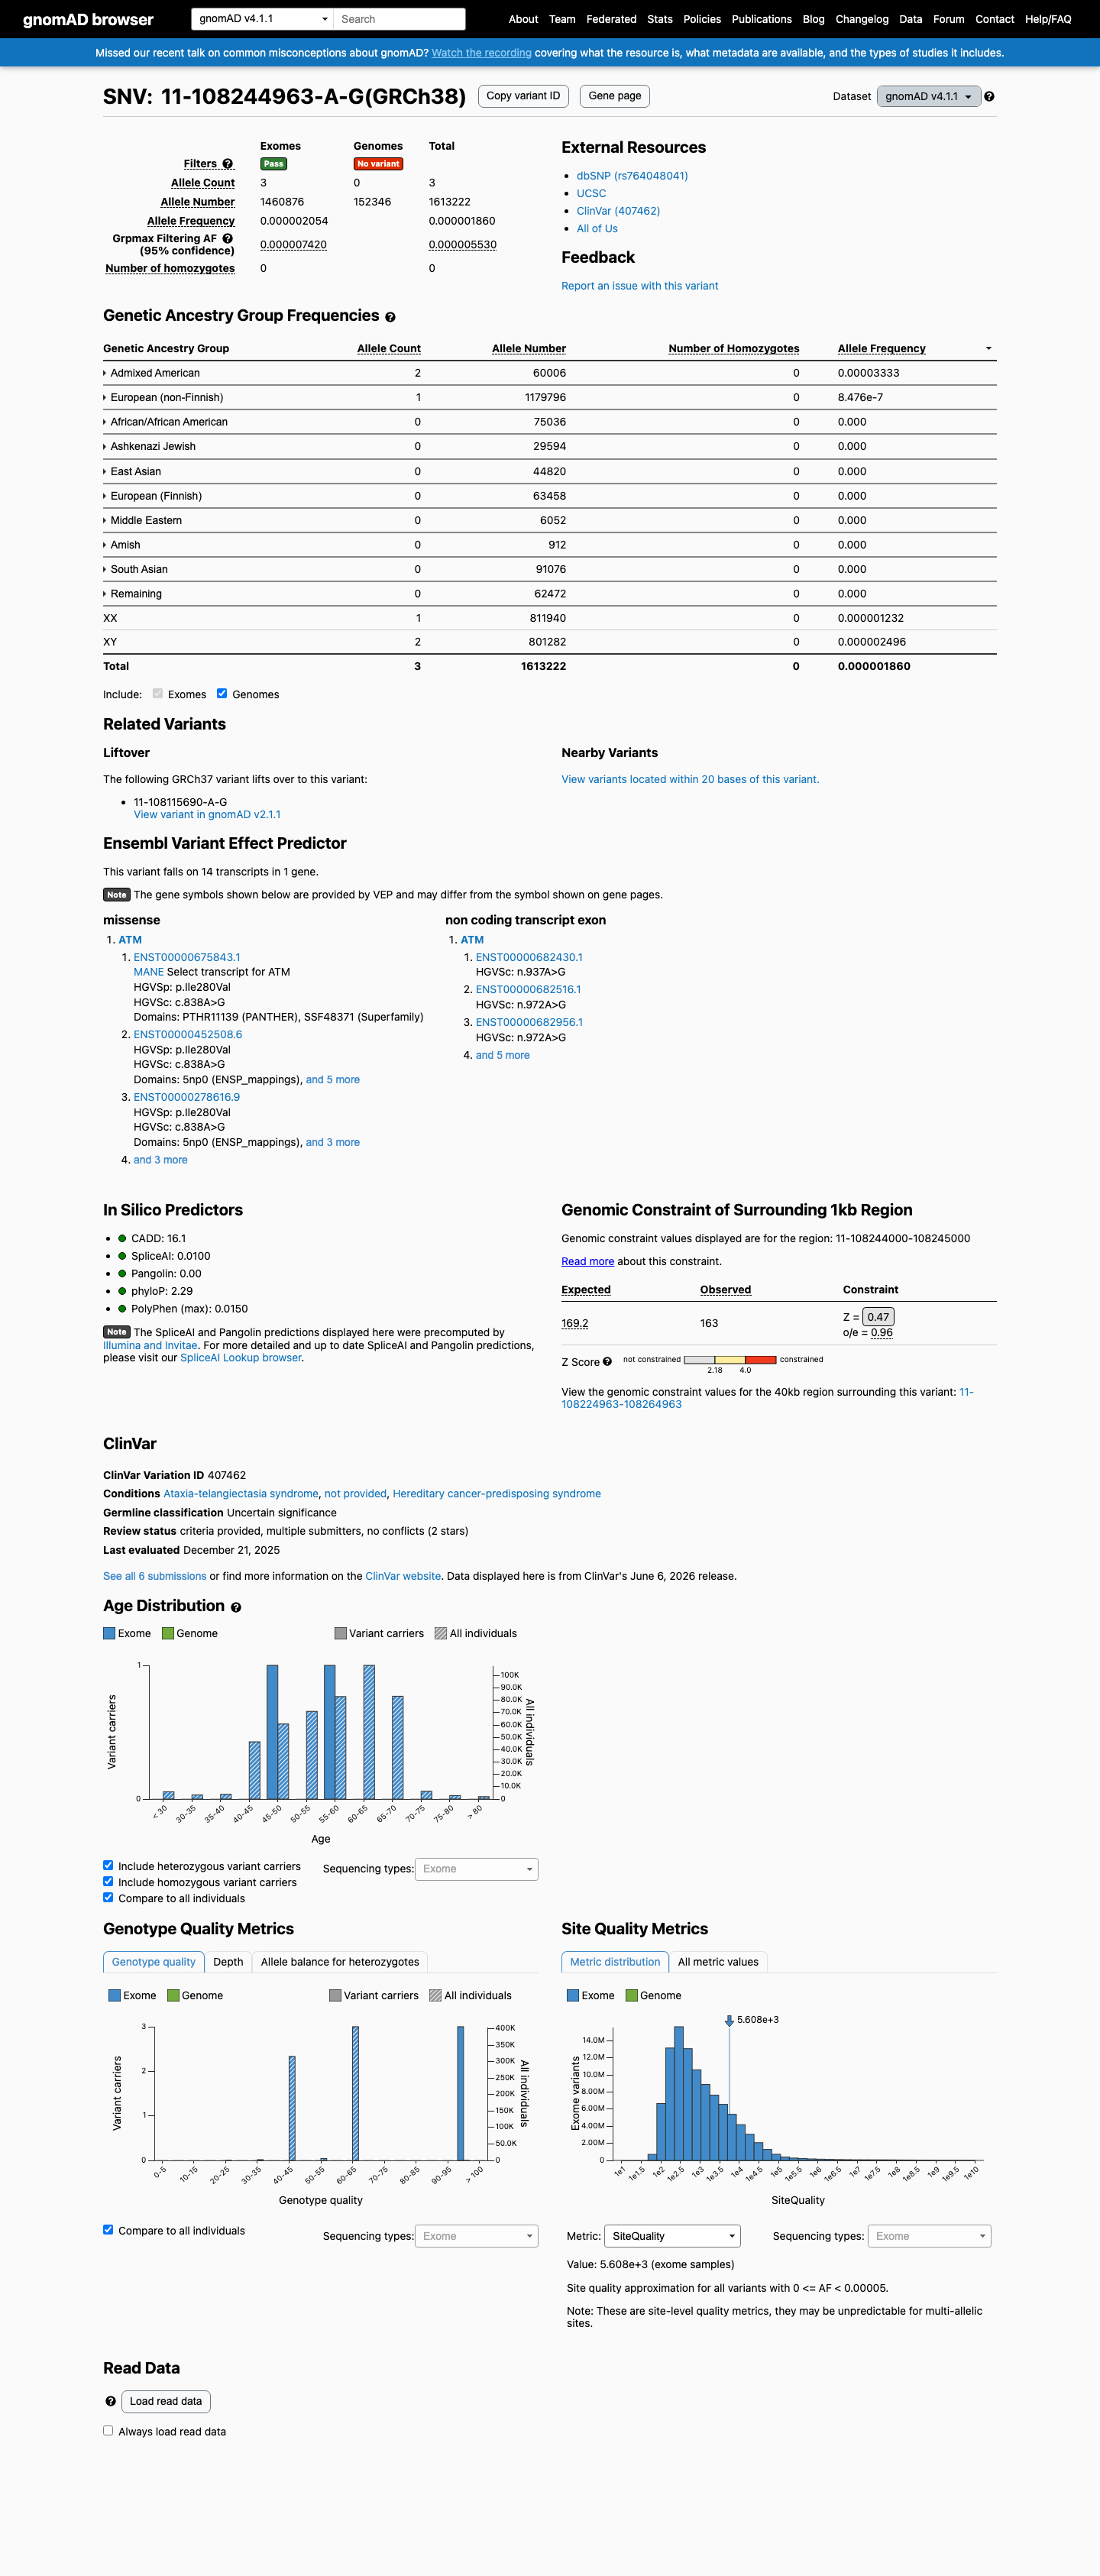
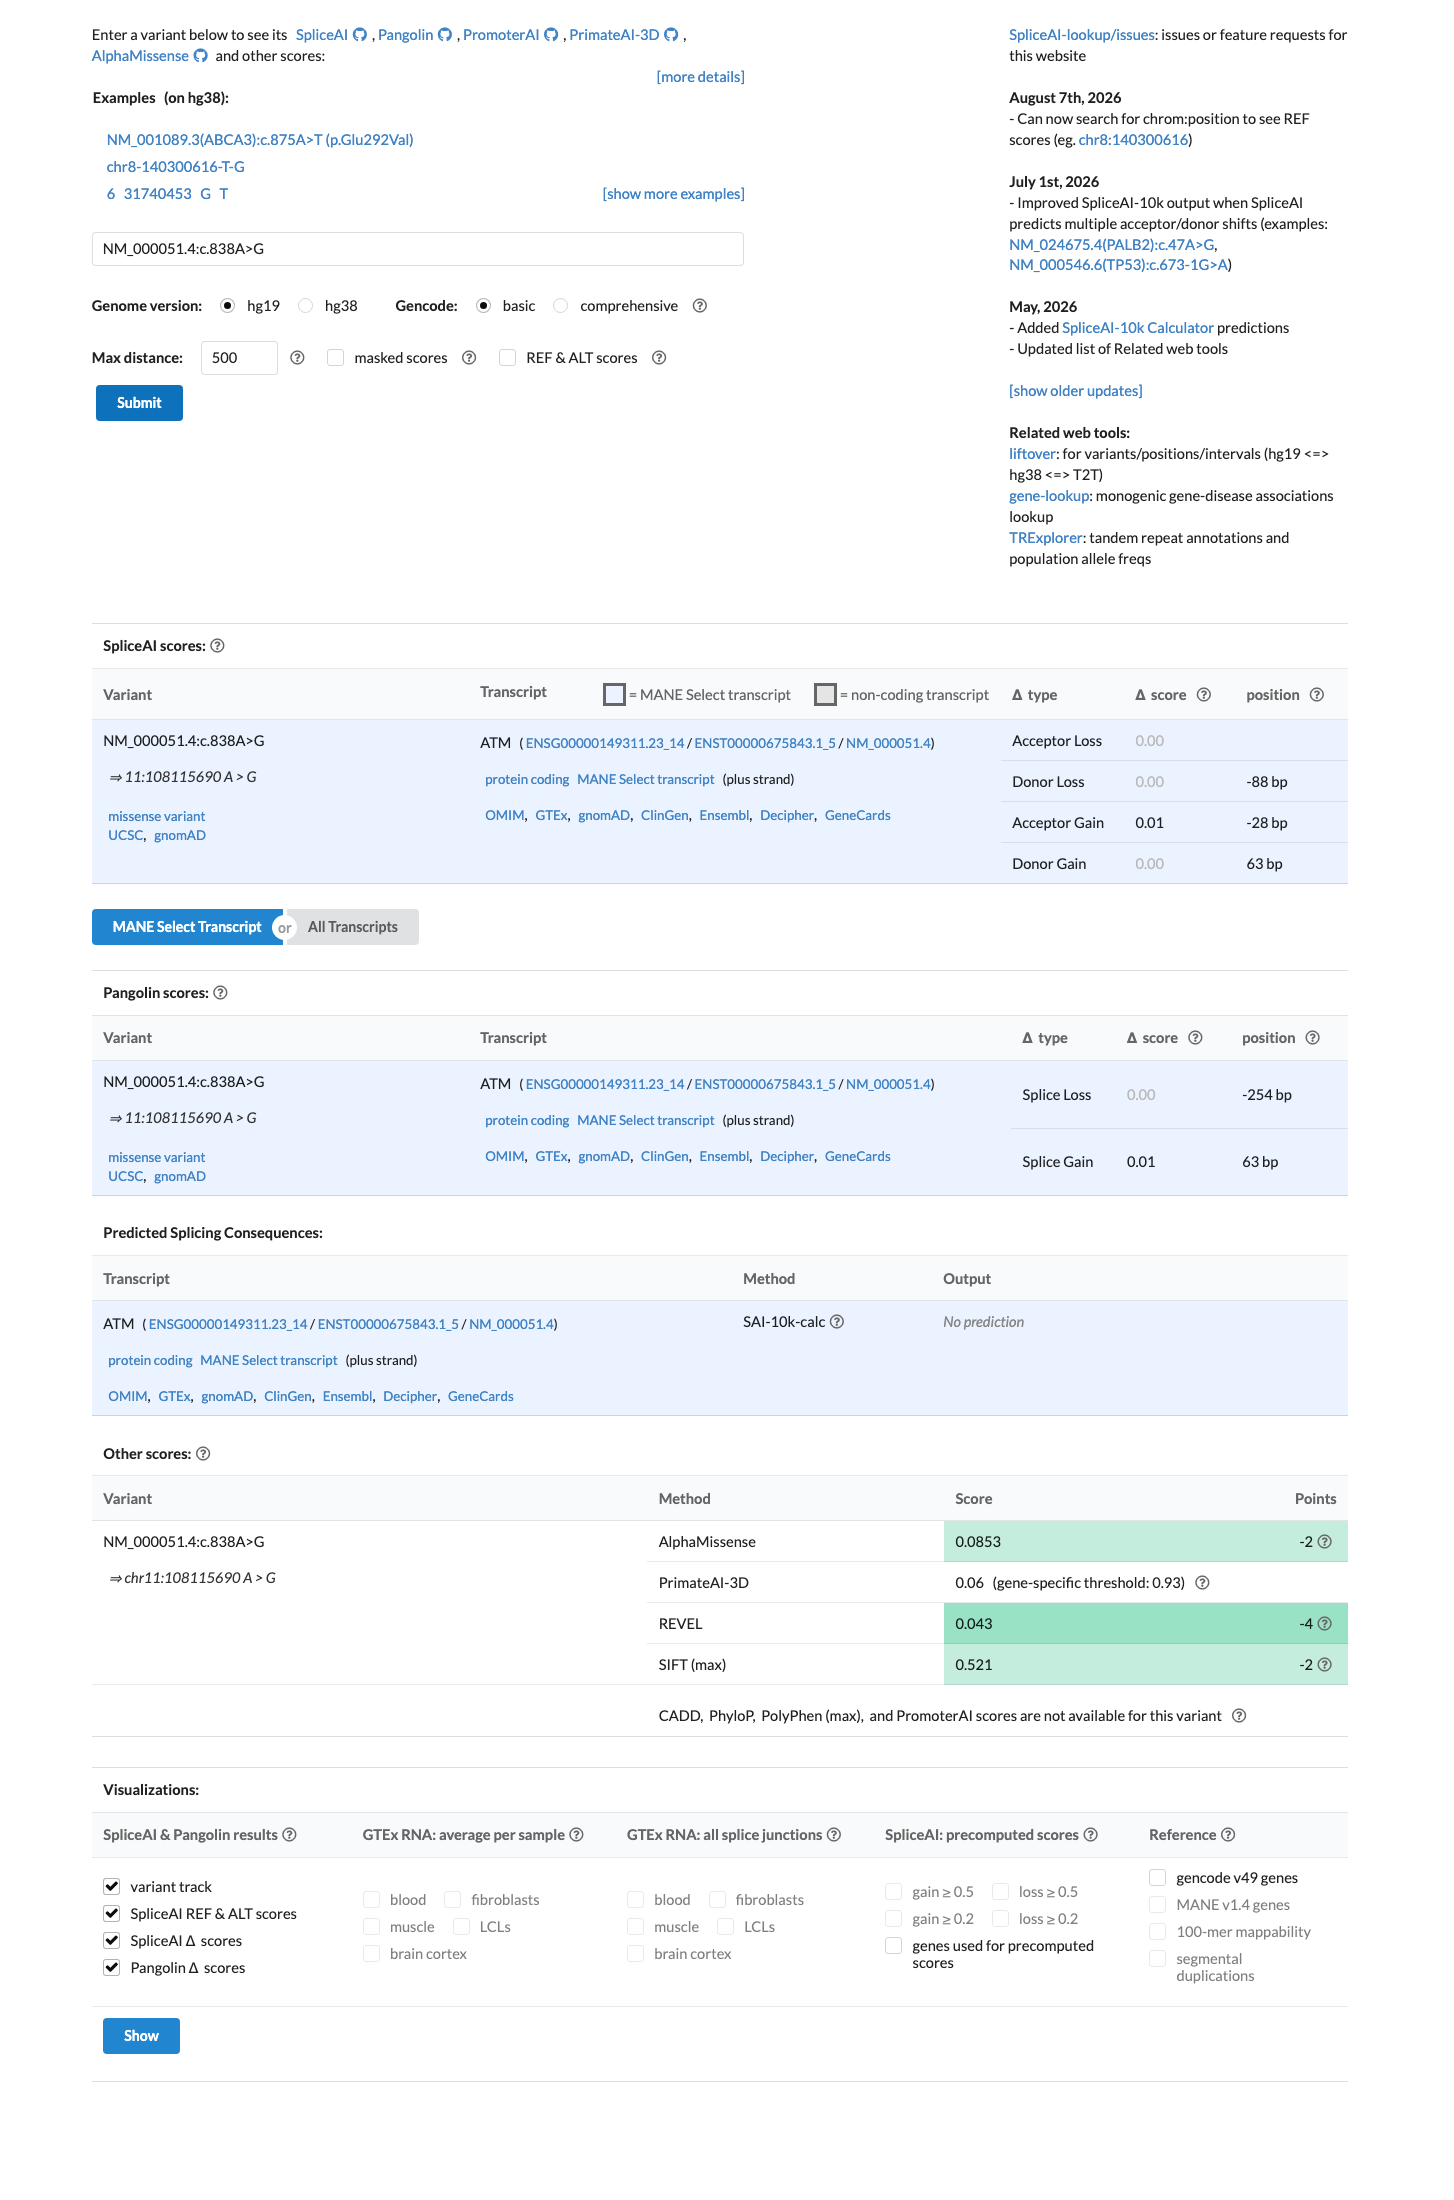
PM2 (Supporting): gnomAD v4.1 total allele frequency 0.000186% (3/1,613,222 alleles) is below the 0.001% threshold, confirming the variant is extremely rare. BP4 (Supporting): REVEL 0.043 is well below the 0.249 missense threshold, indicating a benign-leaning computational effect. Final: PM2 supporting (pathogenic side) combined with BP4 supporting (benign side) satisfies the conflicting-evidence rule, giving VUS (uncertain significance).
PM2 + BP4→VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria
may be applied incorrectly, sources may be misread, and a confident-looking
classification can still be wrong. Double-check every criterion and
its underlying evidence before relying on any call.
Gene diagram
· NM_000051.4 · variants mapped to exon structure
ATMNM_000051.4
Fetching transcript structure from UCSC…
Exons
—
Transcript span
—
Strand
—
Variants mapped
—
Source
UCSC ncbiRefSeqCurated
All variants in ATM—click a row to locate it on the plot · use the link column to open its page
Protein
Location
Classification
Link
Applied criteria · 2 applied · 10 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
✓
PM2supportingPathogenic
Met (supporting): gnomAD v4.1 total allele frequency 0.000186% (3/1,613,222 alleles) is far below the 0.001% threshold.
Assessed · not applied
· 5 not met · 5 not assessed
Pathogenic
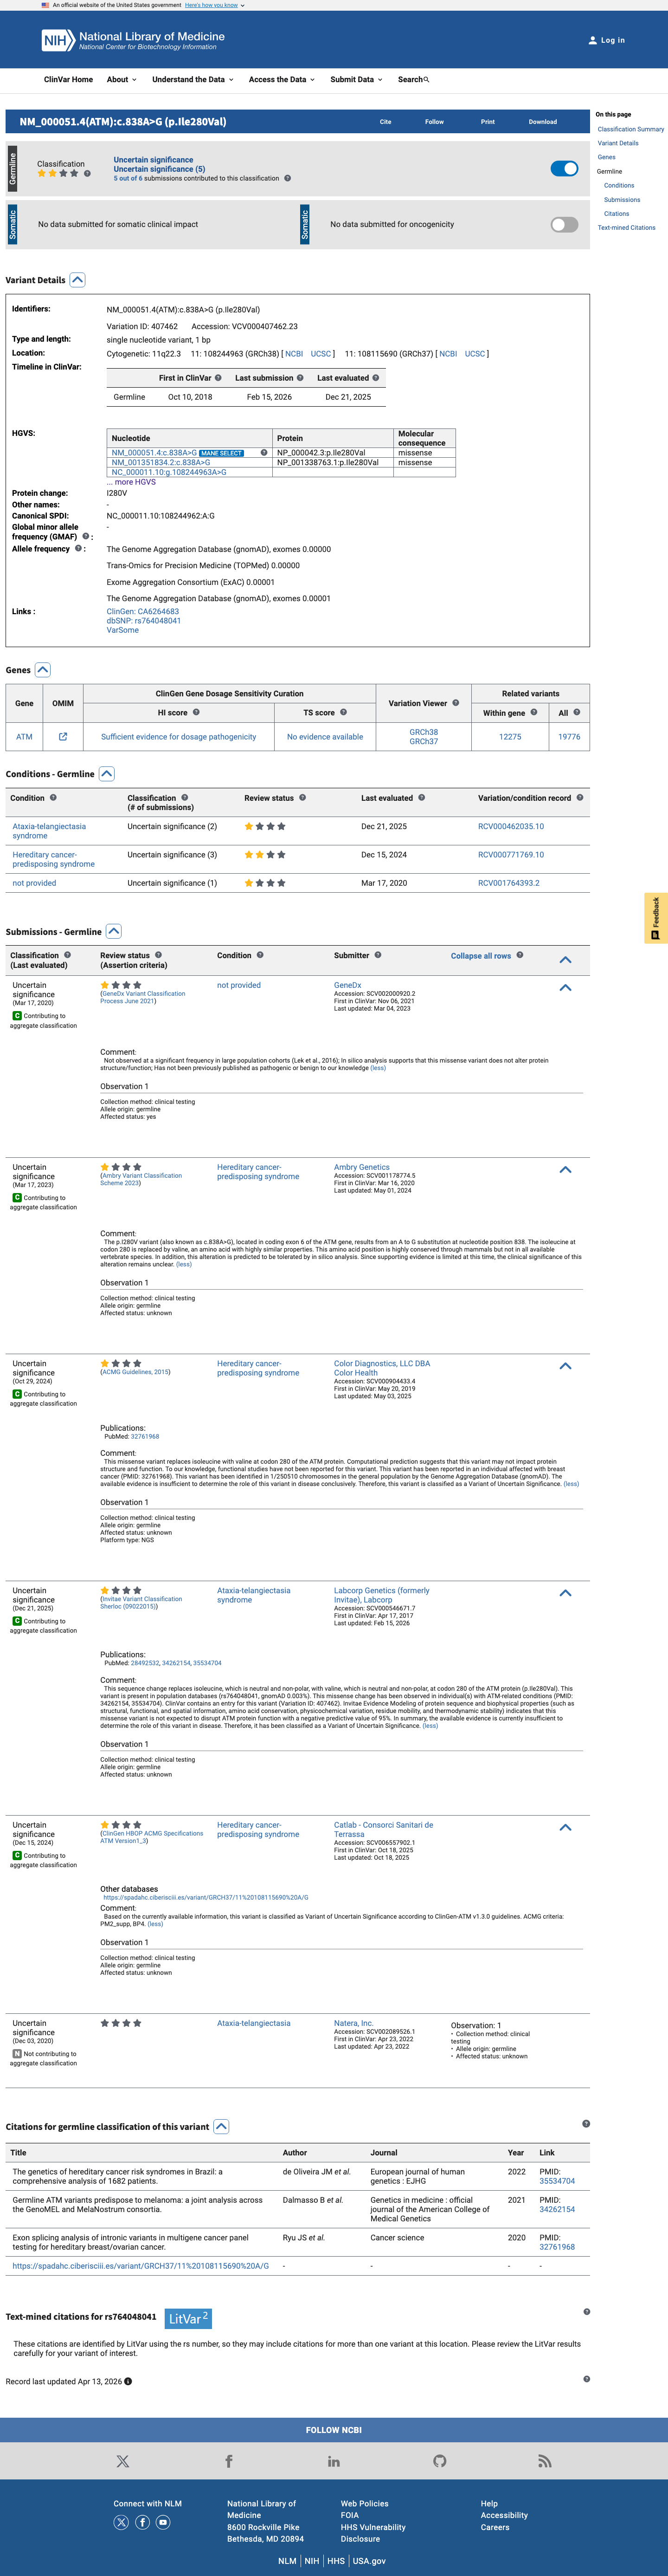
PS1Not met: no established pathogenic or likely pathogenic variant producing p.Ile280Val exists; ClinVar lists only uncertain significance for this variant.
PS3Not assessed: the only functional measurement available (Sun 2025) was equivocal, 'Intermediate', not a confirmed failure to rescue an ATM function.
PS4Not met: no case-control study of this exact variant exists; the only case-control ATM study in the literature does not list c.838A>G.
PM3Not assessed: no ataxia-telangiectasia proband observations of this variant (homozygous or in trans) were available.
PP1Not assessed: no family segregation data for this variant were available (no affected relatives genotyped).
PP3Not met: REVEL 0.043 is far below the 0.7333 threshold, and SpliceAI 0.01 is far below 0.2.
Benign
BA1Not met: grpmax filtering allele frequency 0.000553% is roughly 900-fold below the 0.5% threshold.
BS1Not met: grpmax filtering allele frequency 0.000553% is about 90-fold below the 0.05% threshold.
BS3Not assessed: the only functional measurement available (Sun 2025) was equivocal, 'Intermediate', not a confirmed rescue of function.
BP2Not assessed: no unaffected carriers of this variant with a pathogenic ATM variant in trans were documented.
This variant is present in gnomAD v4.1 (AF= 1.85963e-06; MAF= 0.00019%, 3/1613222 alleles, homozygotes = 0) and has highest observed frequency in the Admixed American population (AF= 3.333e-05; MAF= 0.00333%, 2/60006 alleles, homozygotes = 0); grpmax FAF= 5.53e-06.
v2.1
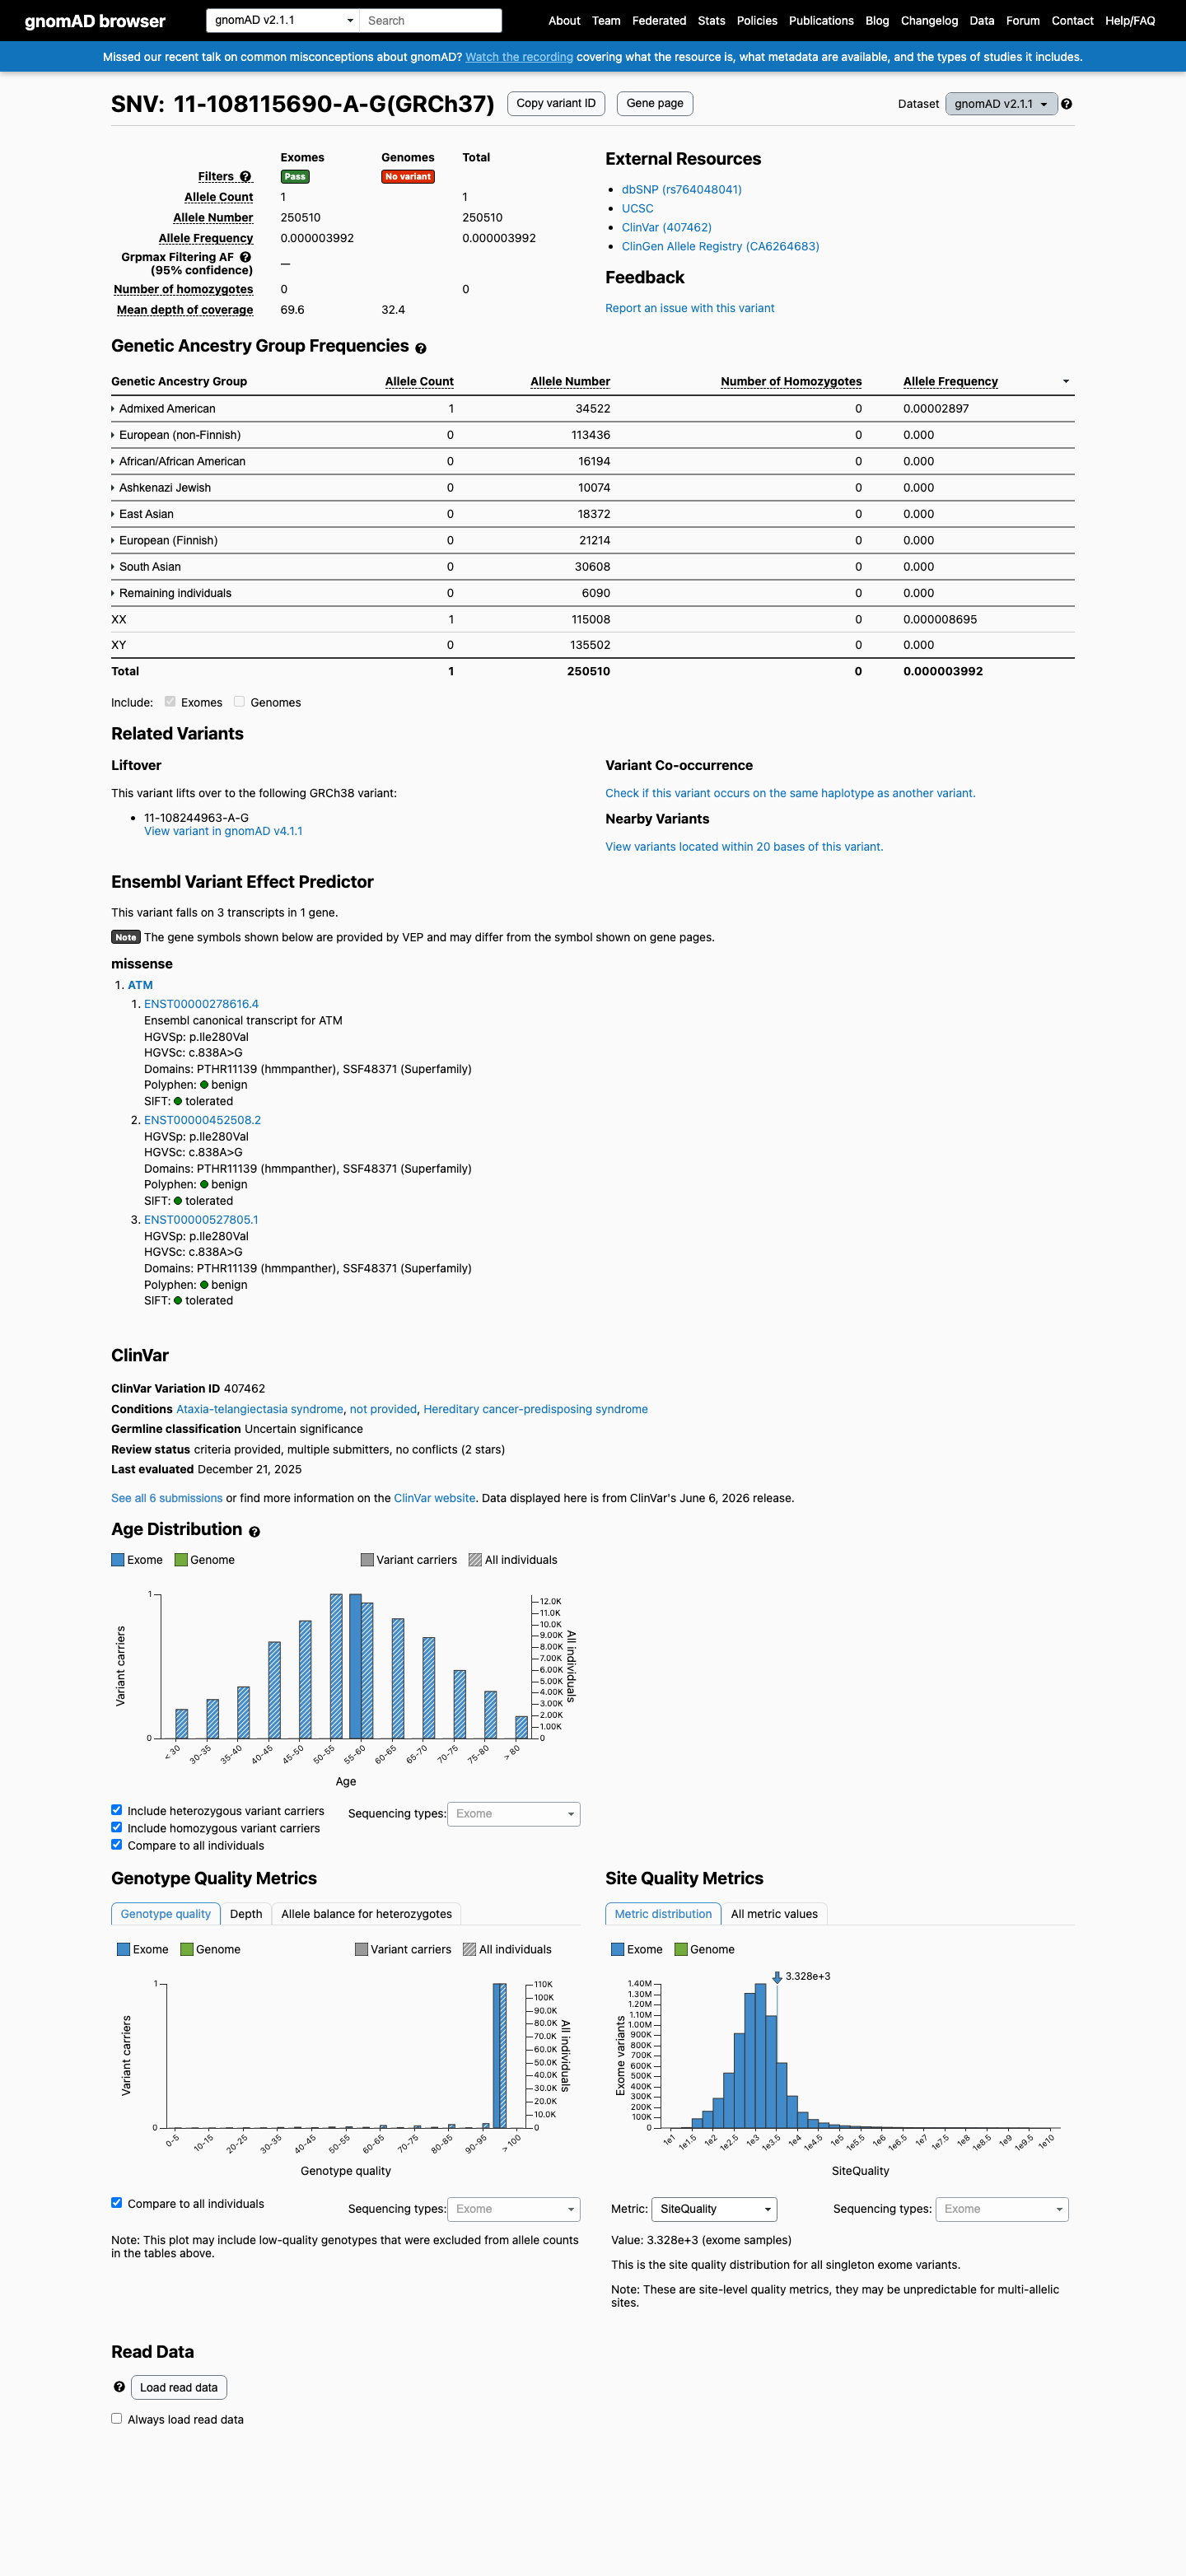
This variant is present in gnomAD v2.1 (AF= 3.99186e-06; MAF= 0.00040%, 1/250510 alleles, homozygotes = 0) and has highest observed frequency in the Admixed American population (AF= 2.8967e-05; MAF= 0.00290%, 1/34522 alleles, homozygotes = 0).
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.00019%
· 3 / 1,613,222
0 hom · FAF 0.00055%
Admixed American
2 / 60,006
0.0033%
European (non-Finnish)
1 / 1,179,796
8.5e-05%
+ 8 not observed (Remaining individuals, European (Finnish), Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American)
gnomAD v2.1
0.0004%
· 1 / 250,510
0 hom
Admixed American
1 / 34,522
0.0029%
+ 7 not observed (African/African American, Ashkenazi Jewish, East Asian, European (Finnish), European (non-Finnish), Remaining individuals, South Asian)
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. ATM, a kinase involved in the DNA damage response, is mutated in various solid and hematologic malignancies.
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 7 further PMIDs triaged but not cited — see Sources & references.
Structured finding pending for this record — see source link.
Applied to
→BP4
supporting
Corroborates BP4: table REVEL 0.043 matches the primary lookup, AlphaMissense/EVE/boostDM are all low, and the measured Classification is 'Intermediate' (not Non-functional, i.e., no damaging signal)
Rule & framework references · cited for criterion definitions, not variant evidence
32761968 ↗Exon splicing analysis of intronic variants in multigene cancer panel testing for hereditary breast/ovarian cancer.
34262154 ↗Germline ATM variants predispose to melanoma: a joint analysis across the GenoMEL and MelaNostrum consortia.
Sources & reference links
9Sources
CSpec VCEP
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 7 PMIDs not cited in assessment
25741868 ↗Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.CLINVAR
24418350 ↗EFNS/ENS Consensus on the diagnosis and management of chronic ataxias in adulthood.CLINVAR
25394175 ↗A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment.CLINVAR
35534704 ↗The genetics of hereditary cancer risk syndromes in Brazil: a comprehensive analysis of 1682 patients.CLINVAR
20050888 ↗EFNS guidelines on the molecular diagnosis of ataxias and spastic paraplegias.CLINVAR