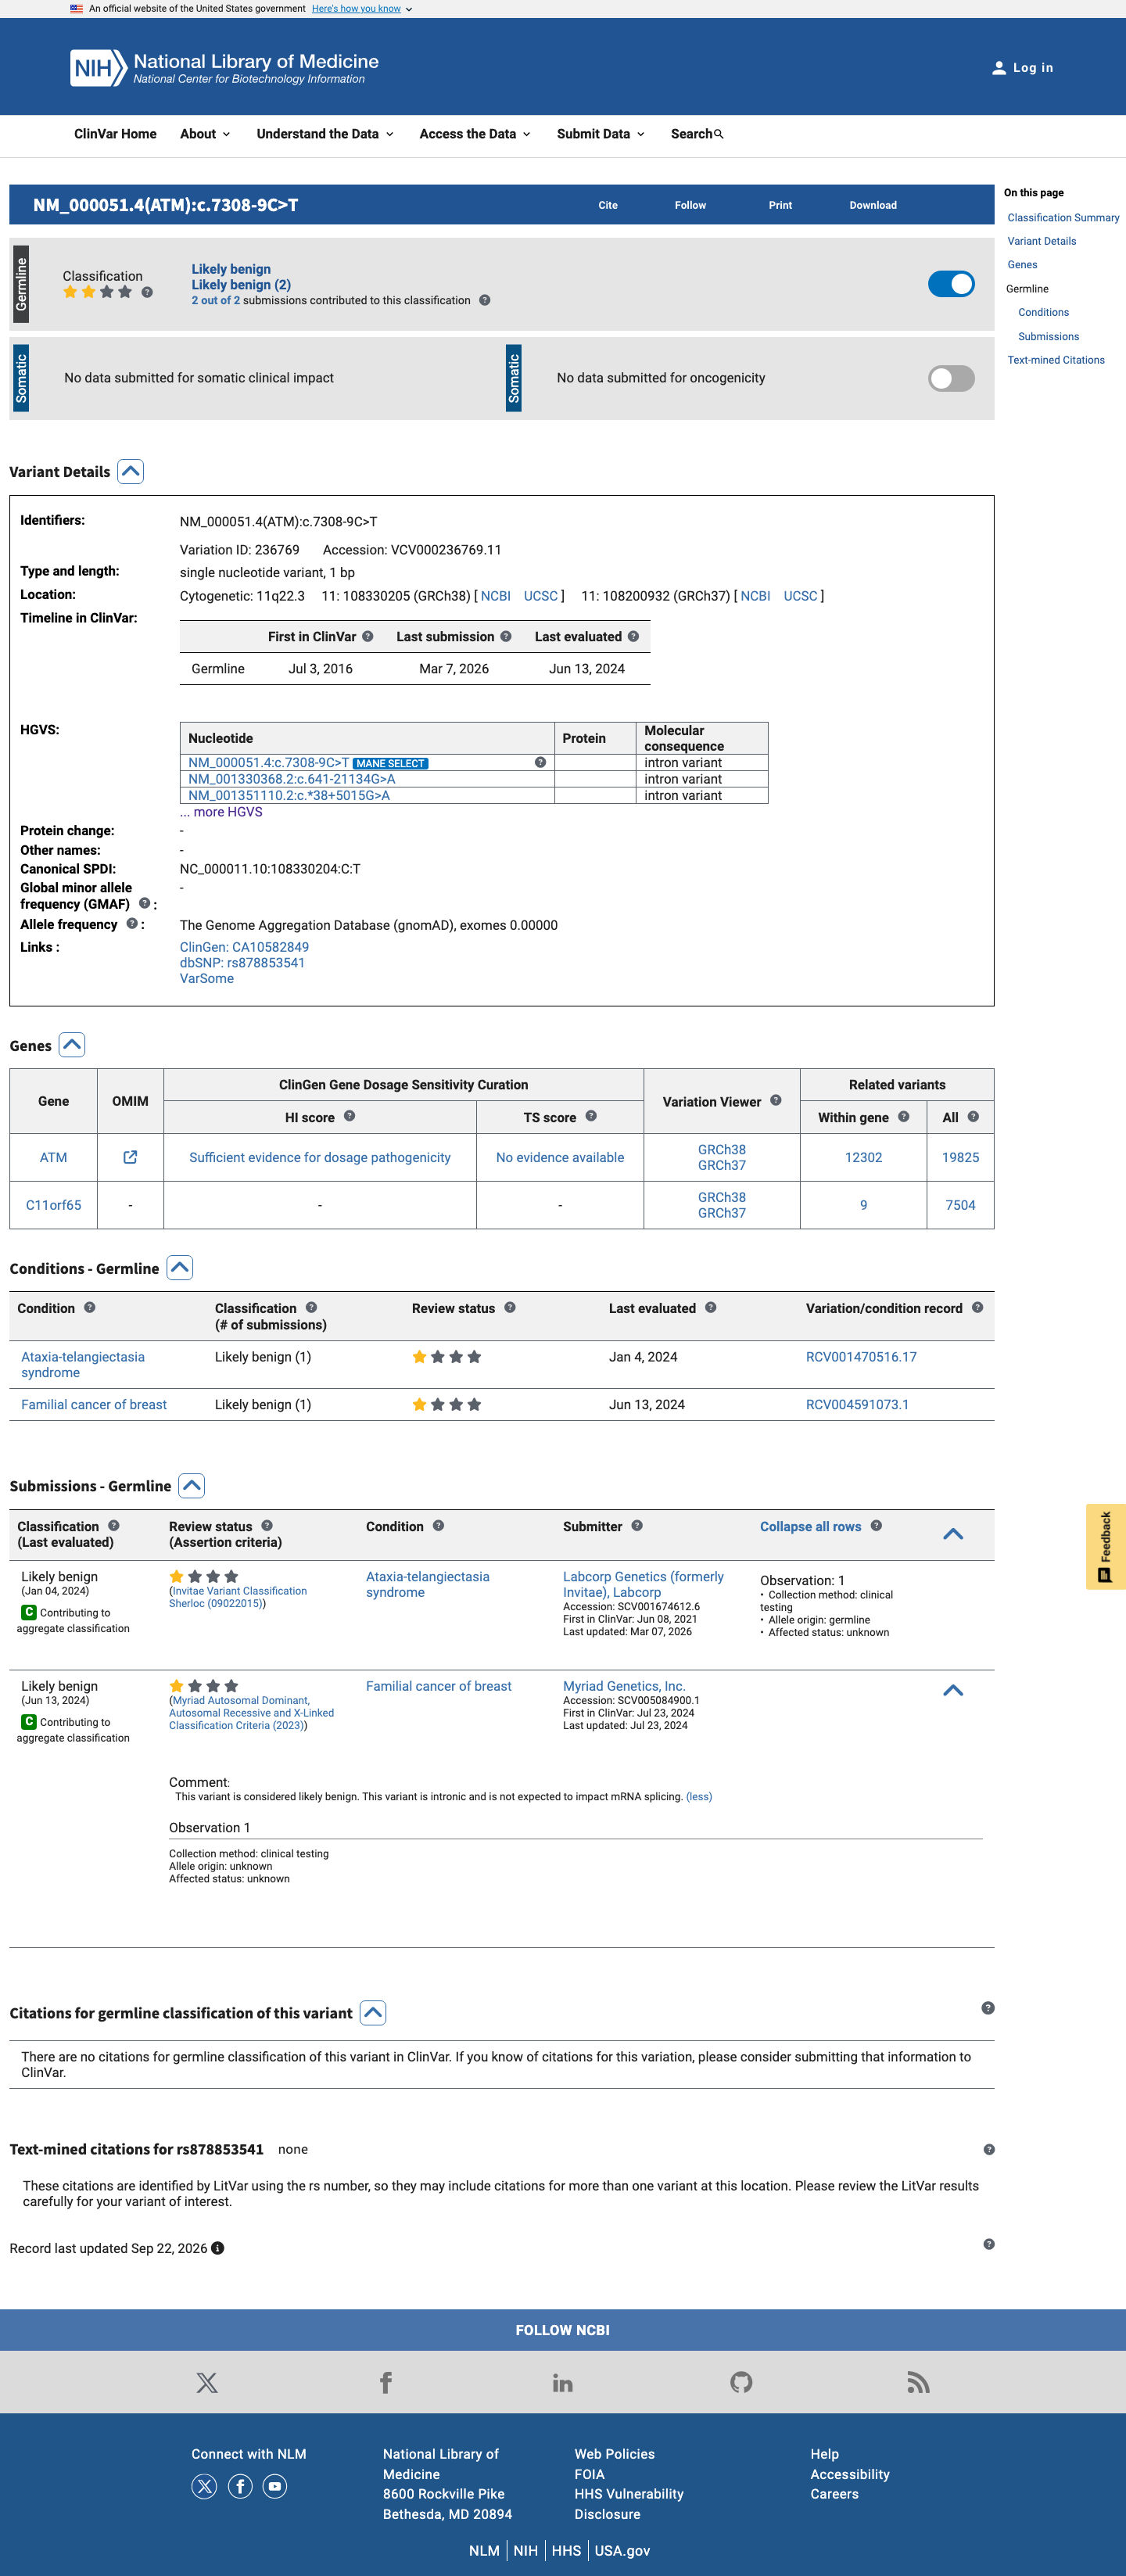
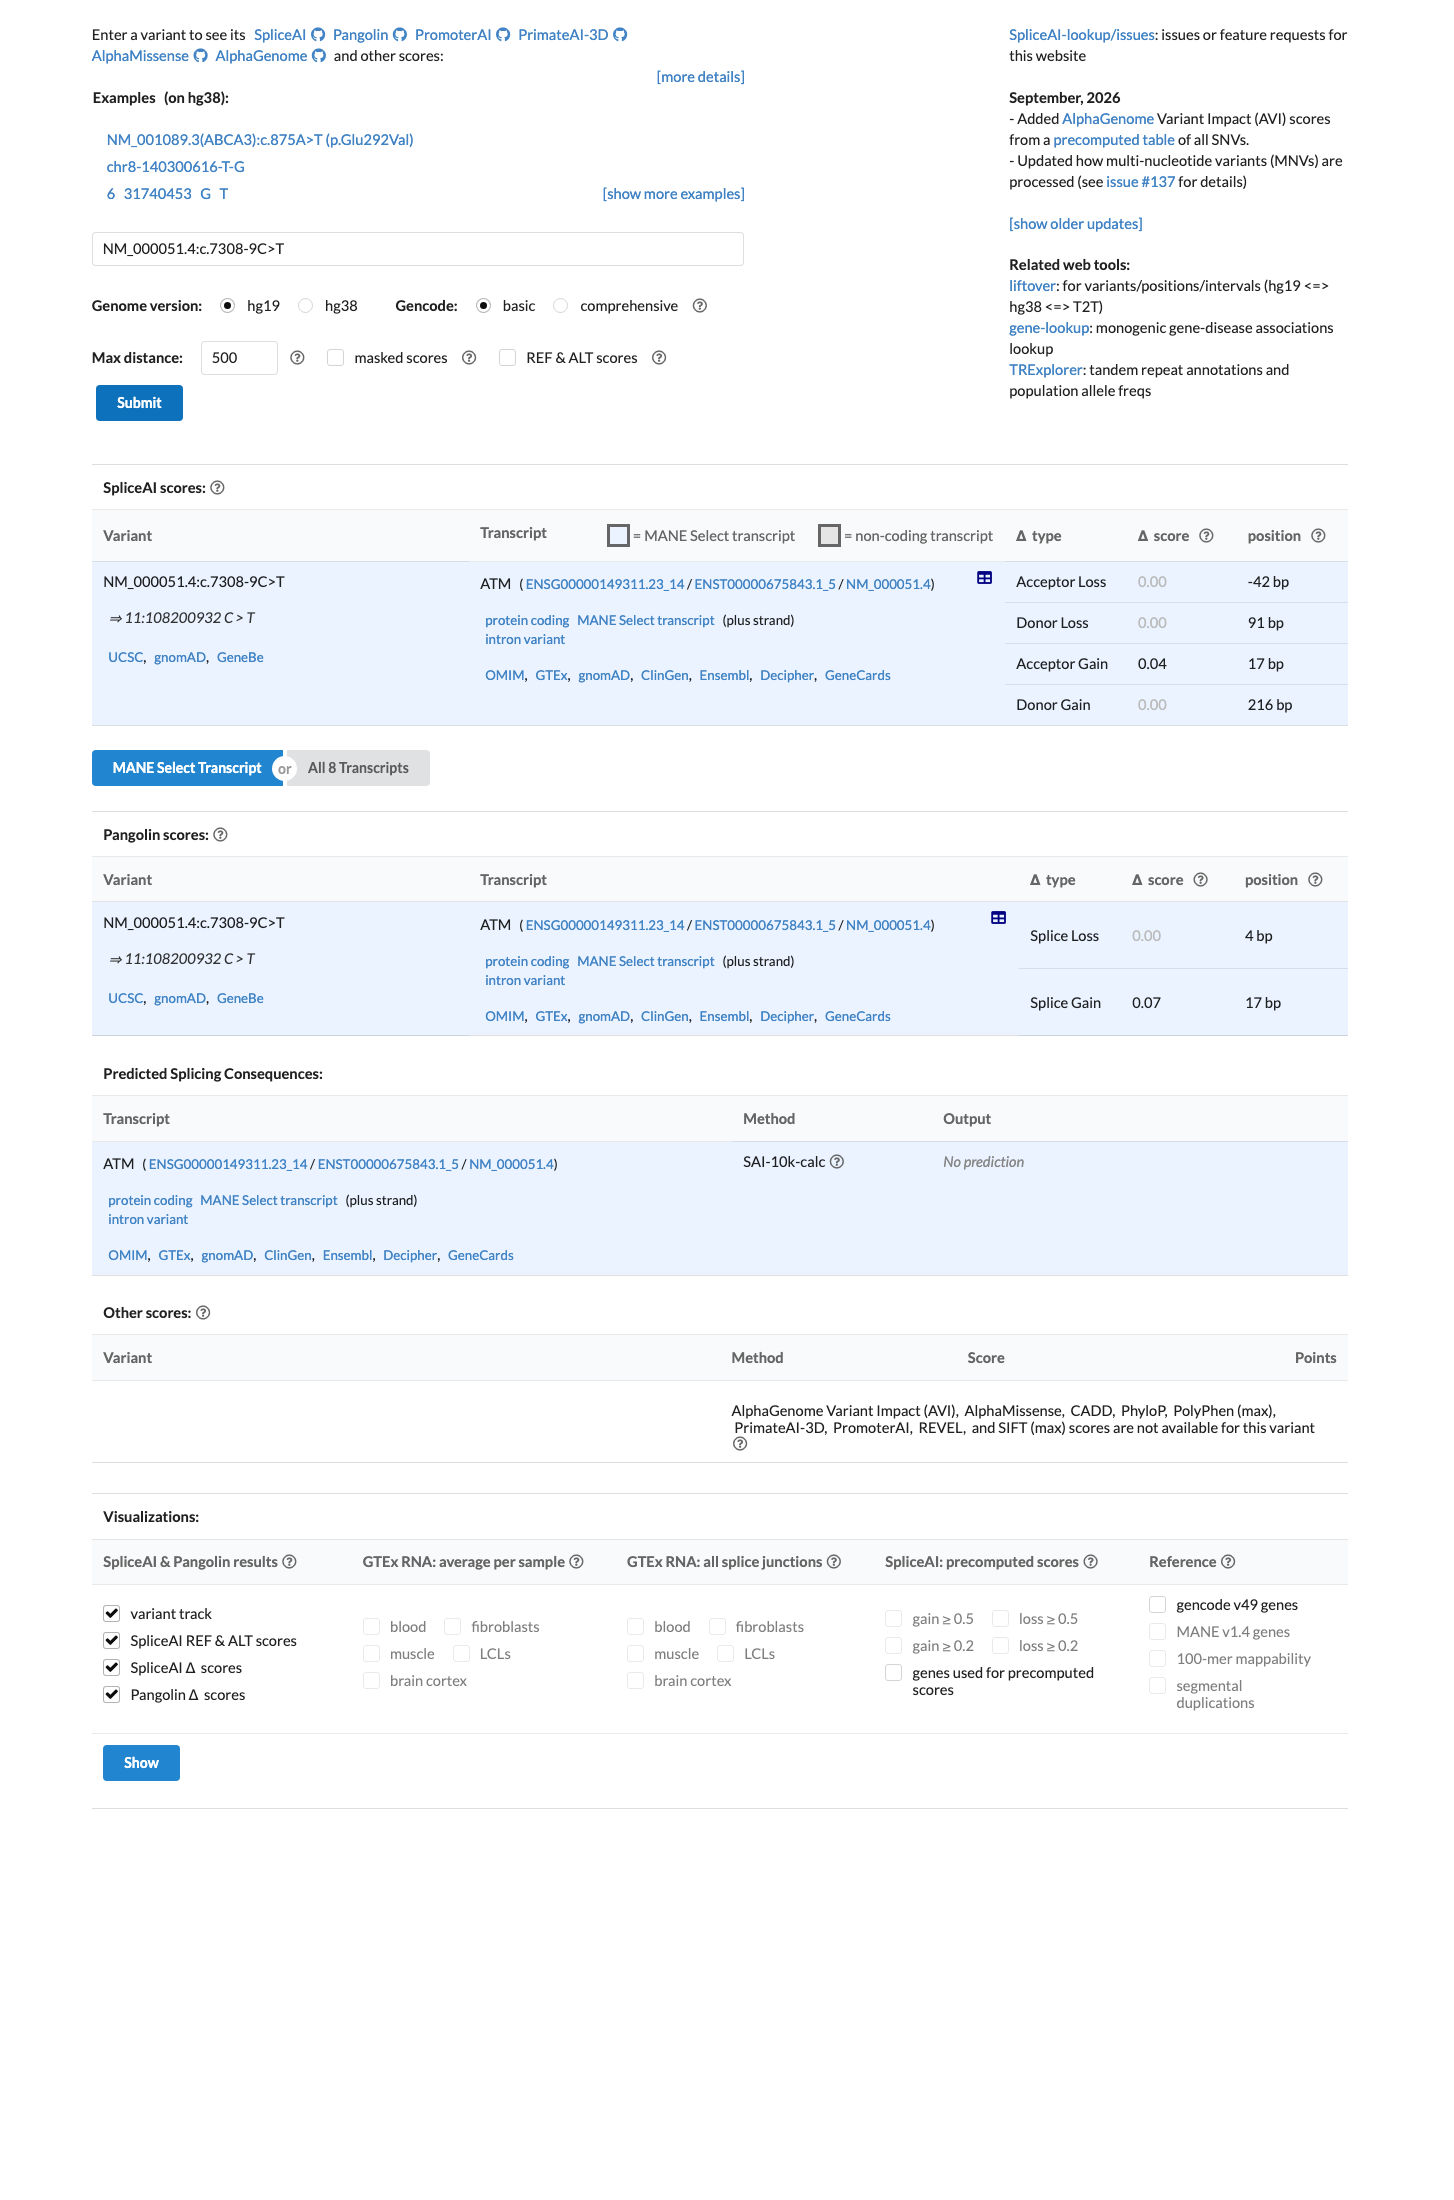
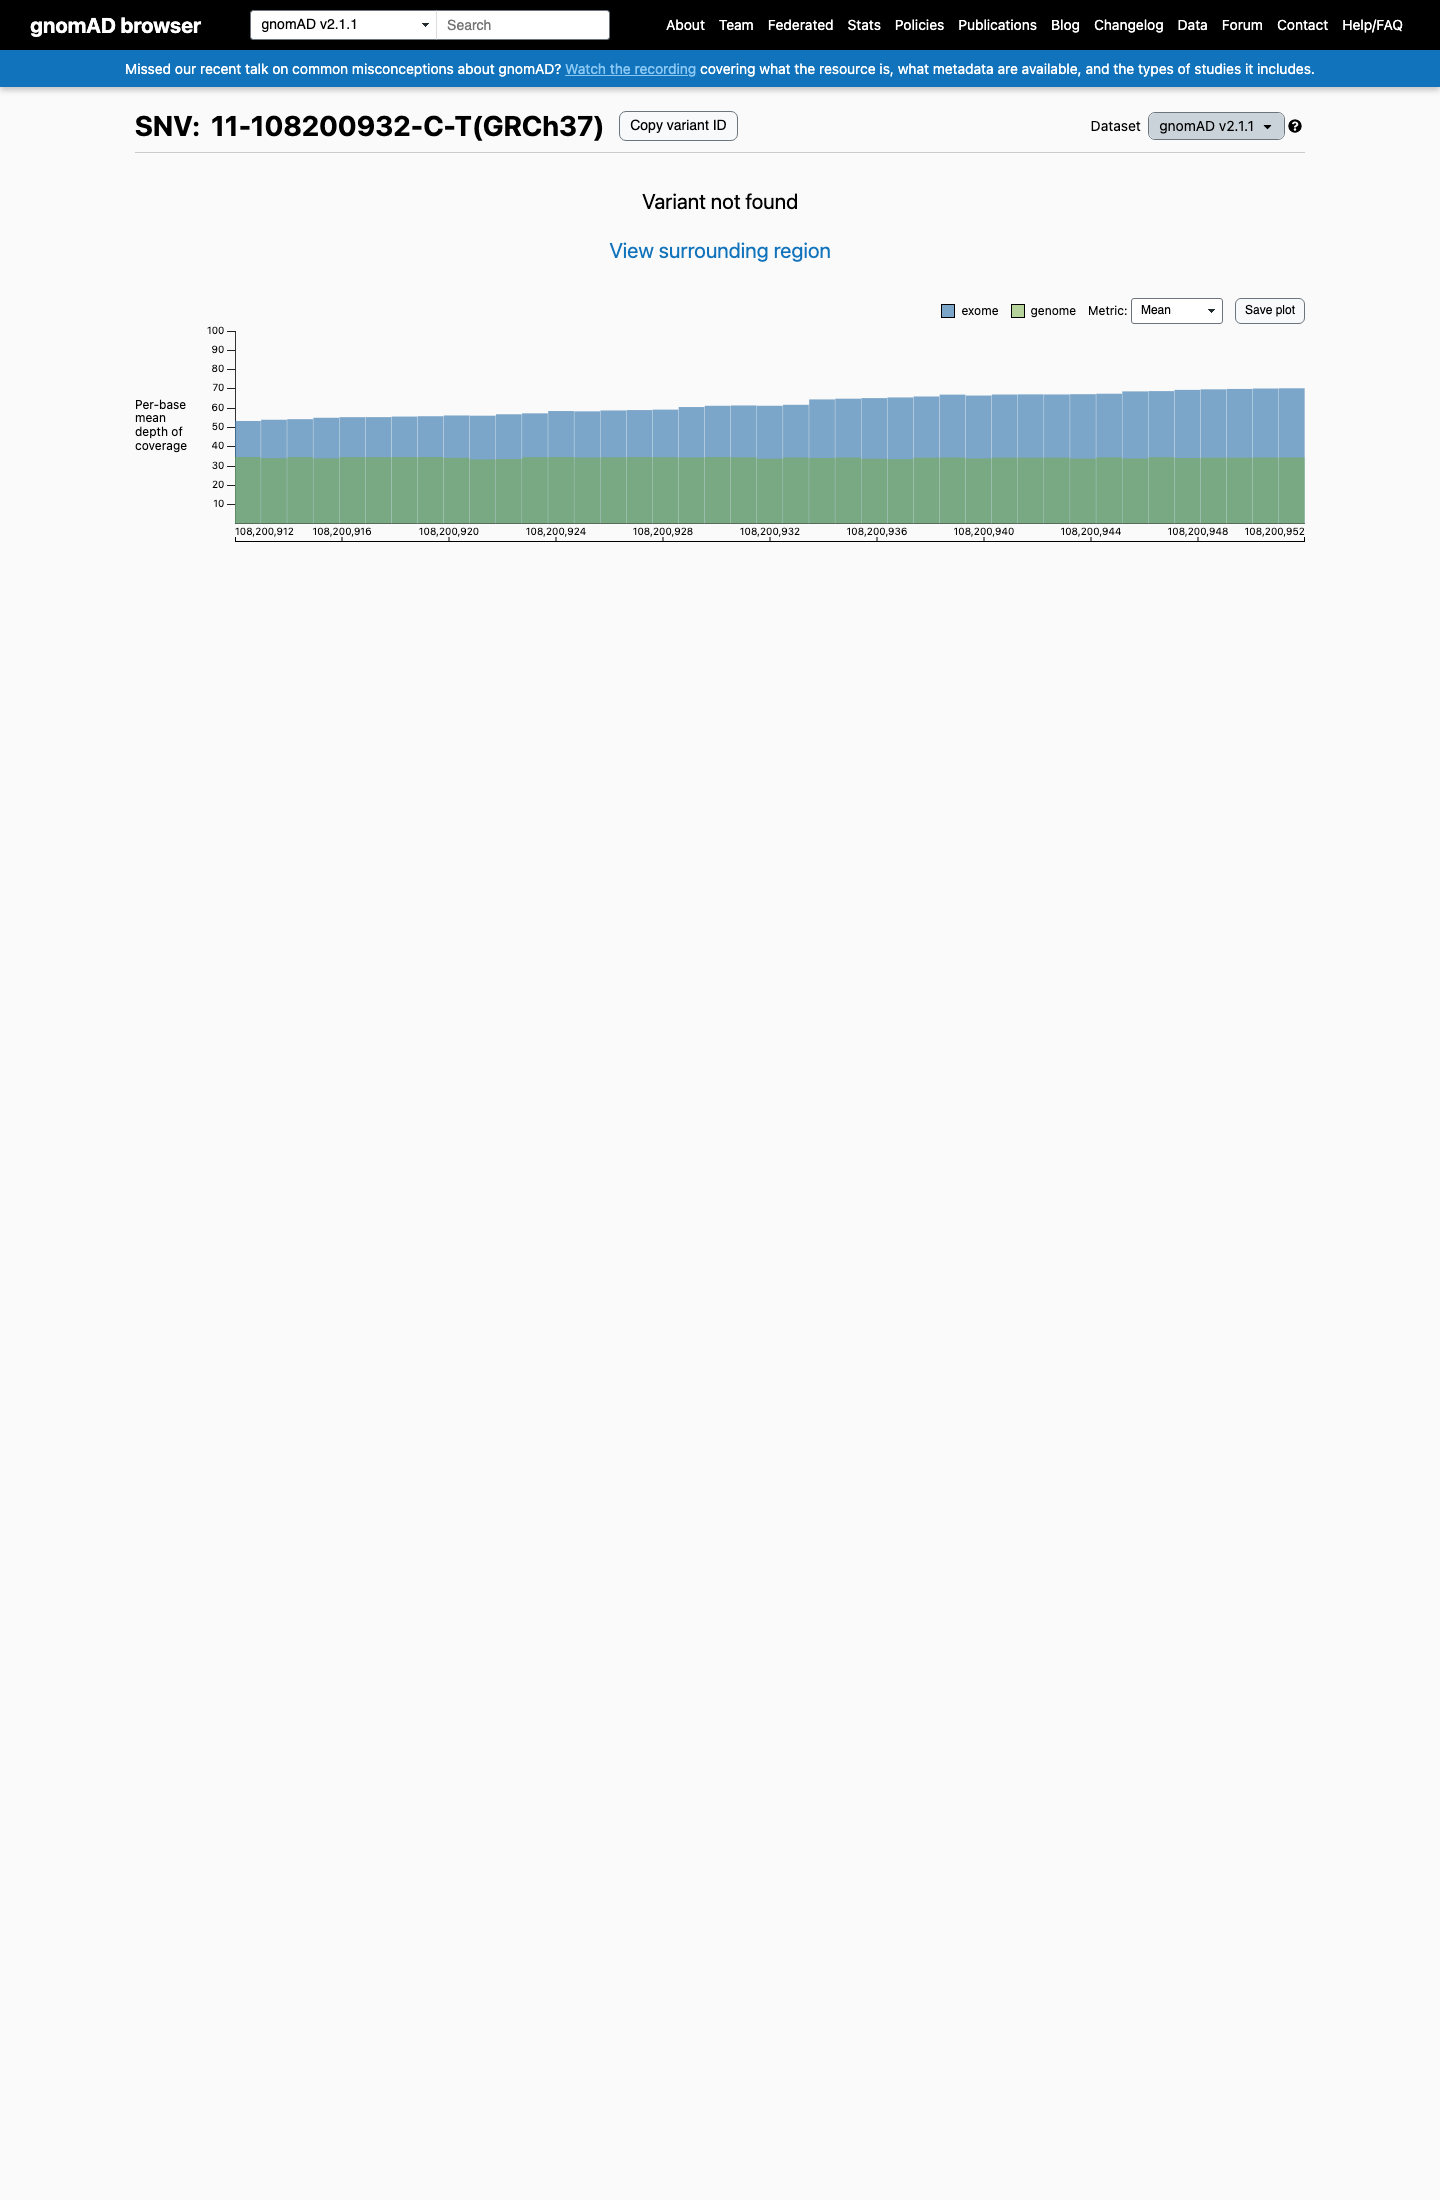
Consequence/LOF assessment for ATM NM_000051.4:c.7308-9C>T, an intronic substitution 9 nt upstream of the exon 50 acceptor site (11:108330205C>T GRCh38 / 11:108200932C>T GRCh37), predicted protein consequence NP_000042.3:p.?. PVS1 is not met: the variant is not a null variant under the governing ClinGen HBOP ATM VCEP v1.5/v1.6 PVS1 criterion (nonsense, frameshift, canonical +/-1 or 2 splice site, initiation codon, single/multi-exon deletion); it lies outside the canonical +/-1,2 dinucleotide and SpliceAI max delta is 0.039, so no predicted splice defect exists and the observed-defect PVS1_Variable(RNA) route cannot be entered without RNA data. No strength tier is assigned. PM4 is not met: the ATM VCEP restricts PM4 to stop-loss variants, and this intronic substitution neither creates a stop-loss nor changes protein length. BP3 is not applicable: the ATM VCEP specifies BP3 as 'Not applicable - do not use', and the variant is not an in-frame indel in a repeat region in any case. No consequence-level evidence supporting a pathogenic direction was found for this variant in any source consulted; the ClinVar consensus (Likely benign, 2 submitters, no expert panel) was not used to infer or contradict any criterion in this group. This group's five criteria are resolved mainly by the gene-specific ATM HBOP VCEP specification (CSPEC v1.6), which is the governing framework: PM1, PP2 and BP1 are explicitly listed as Not applicable for ATM with stated reasons (benign and pathogenic variants co-occur within the same domains and hotspots are ill-defined; ATM has no defined low rate of benign missense variation; pathogenic missense variants are known for ATM). These three are recorded as not_applicable rather than unmet, so that a downstream reviewer does not read them as absent evidence. Only PS1 and PM5 remain genuinely evaluable, and both fail. PS1 cannot be entered: the ATM PS1 splicing table requires the variant under assessment to carry its own baseline predictive code and to share a precisely matching predicted splice event with a P/LP reference variant of equal or lower prediction strength, and no P/LP reference variant exists at the acceptor c.7308-9 position (ClinVar 236769, the variant itself, is Likely benign with 2 submissions and no expert panel; the nearest P/LP changes c.7308-2A>C and c.7308-1G>C are canonical acceptor variants at different positions). The VUA also has no predicted splice event (SpliceAI max delta 0.039, below the >= 0.2 PP3 splicing cut-off), so the prerequisite of a matching event cannot be met. PM5 is likewise not met. The VCEP's PM5 is a truncation rule granting only PM5_Supporting to NMD-prone truncating variants with PVS1 at Very Strong and a premature termination codon upstream of p.Leu3048, with the splice path restricted to observed (not predicted) effects on high-quality RNA data; it explicitly states the path is not applicable for predicted splice impact without RNA data and forbids use for missense changes. c.7308-9C>T is a non-truncating intronic substitution with no amino-acid change, no PVS1, no RNA data and no same-residue comparator (pm5_candidates.json found = false). Cross-criterion consistency note: for a non-missense, non-truncating intronic variant (p.?) there is no residue to place in a domain or hotspot, so PS1, PM1, PM5, PP2 and BP1 are all unavailable on this variant's own terms as well as under the VCEP's gene-level decisions - the residue/codon/domain axis contributes no pathogenic evidence for this variant. No functional assay evidence exists for NM_000051.4:c.7308-9C>T (intron 49, -9 from the exon 50 acceptor): the variant is absent from both VCEP-declared functional sources (clingen_hbop_atm_supplementary_tables_1_and_2_v1.xlsx, an assay-strength calibration table only; and Sun et al. 2025 PMID 40580951 Table S1, whose intronic coverage stops at offsets -5..-1/+1..+5). Because no readout (damaging or rescuing) exists, neither PS3 nor BS3 is met, and the ATM HBOP VCEP v1.6 strength ladder for either criterion could not be applied; PS3_Strong is expressly disallowed under this framework. PM3 and BP2 for ATM are not single-observation codes: under the ClinGen HBOP ATM VCEP PM3/BP2 points table they are direction-opposed tallies of per-individual observations that convert to a strength (PM3_Supporting >= 1, PM3 = 2, PM3_Strong = 4, PM3_VeryStrong >= 8; BP2_Supporting <= -1, BP2_Moderate = -2, BP2_Strong <= -4). Both halves of that table were applied and both totalled zero points: no unrelated A-T proband carrying c.7308-9C>T (any phenotype class, any phase), and no unaffected non-A-T adult carrying it with a P/LP ATM variant (in trans or homozygous, laboratory or database setting). The table's applicability gate is met rather than failed: gnomAD v4.1 total AF 6.2032e-07 (1/1,612,070 alleles, 0 homozygotes), absent from gnomAD v2.1 and gnomAD-Canada, well below the table's 0.01% ceiling and below the ATM VCEP PM2_supporting ceiling of 0.001%. The null result is therefore a genuine absence of proband-level observations, not a frequency-based disqualification. Variant class is not a limitation for this table - the VCEP states the PM3/BP2 approach applies regardless of variant class - so an intronic variant 9 bp upstream of the exon 50 acceptor site (NC_000011.9:g.108200932C>T / chr11:108330205C>T) was fully eligible for consideration; SpliceAI max delta 0.039 means the variant is not predicted to disrupt splicing, which removes any concern about applying the points route. Evidence searched and found negative: ClinVar variation 236769 (two Likely benign clinical-laboratory submissions, zero expert-panel submissions, zero criterion-level leads, no validated PMIDs); gnomAD v2.1/v4.1/gnoMAD-Canada (1 heterozygous allele in total, 0 homozygotes); the ATM PM3/BP2 v1.5 and v1.6 tables (points rules only, no per-variant entry); and the 6-PMID literature packet (no ATM mention in any paper). Neither criterion is met, and neither is directionally supported: PM3 records 0 of the >= 1 points required, BP2 records 0 of the <= -1 points required. The variant's overall benign-leaning ClinVar status is a separate assertion evaluated by other criteria groups and was not used to reason towards BP2, per the instruction not to infer a criterion from another source's final classification. c.7308-9C>T is an intronic acceptor-region substitution (intron 49, 9 nucleotides from the exon 50 acceptor site), so PP3 and BP4 were evaluated solely through the SpliceAI splice path and REVEL was not consulted. SpliceAI maximum delta 0.039 satisfies the ATM VCEP v1.6 BP4 no-splice-impact threshold of <=0.1 (BP4 met, supporting) but falls below the VCEP PP3 threshold of >=0.2 (PP3 not met); the same single prediction is not counted twice across the two criteria. BP7 is not applicable because the variant is neither synonymous nor a VCEP-defined deep intronic variant (it sits at -9, inside the -21 acceptor boundary). The exact variant was searched in the governing VCEP full-text sources; Suppl_TableS1_PMID 40580951.xlsx contains no row for c.7308-9C>T and therefore pre-assigns no PP3/BP4/BP7 code, and the VCEP-approved functional-assay table carries no computational codes. Population-frequency group (BA1, BS1, BS2, PM2) adjudicated under the governing ClinGen HBOP ATM VCEP v1.6 specification (cspec doc 639508985), which is instrument-specific and takes precedence over generic ACMG/AMP defaults; generic thresholds were therefore not used. Variant context: ATM NM_000051.4:c.7308-9C>T (NC_000011.10:g.108330205C>T), an intronic (exon 49i) substitution, extremely rare in every population source queried. gnomAD v4.1: total AF 6.2032e-07 (1/1,612,070 alleles, 0 homozygotes), exome-only 6.85e-07 (1/1,459,862); highest subpopulation European (non-Finnish) 8.4857e-07 (1/1,178,460, 0 homozygotes); all other subpopulations 0. gnomAD v2.1, gnomAD-Canada v1.0, and both non-cancer subsets (v2.1 non-cancer exomes, v3.1 non-cancer genomes) report the variant absent. BA1 (VCEP: Grpmax Filtering AF >0.5%) is not met - the highest subpopulation AF is ~5,900-fold below threshold. BS1 (VCEP: Grpmax Filtering AF >0.05%) is not met - ~590-fold below threshold. Both verdicts are robust to the un-captured grpmax filtering AF. BS2 is not applicable: the ATM HBOP VCEP v1.6 explicitly withholds BS2 and provides no strength rules for it; independently, no homozygote is observed in any source, so no BS2-type observation exists. PM2 is met at Supporting strength: the highest gnomAD subpopulation frequency (NFE 8.4857e-07) is below the VCEP's <=0.001% cutoff, and the ancillary n=1-in-a-single-subpopulation/absent-elsewhere exception is also satisfied; only a Supporting-strength PM2 rule is provided by the specification. The ATM HBOP VCEP v1.6 specifies no non-cancer or exome-only population dataset for PM2/BA1/BS1/BS2, so the all-comers GNOMAD_V2_1/GNOMAD_V4_1 entries were the primary source; the GNOMAD_V2_1_NON_CANCER and GNOMAD_V3_1_NON_CANCER subsets were additionally checked and are concordant (absent). Net population-group contribution to the final classification: PM2_Supporting only; no benign population criteria are met.
ATM encodes a kinase that acts as a master controller of the cellular DNA damage response: it detects double-strand breaks and coordinates DNA repair, cell cycle arrest, or apoptosis through downstream targets including p53, BRCA1, and CHK2. Loss-of-function mutations in both copies of ATM cause ataxia telangiectasia, an autosomal recessive disorder characterized by neurological problems, immune deficiency, and cancer predisposition. ATM functions as a tumor suppressor: individuals with ataxia telangiectasia are prone to childhood lymphomas, leukemias, and breast cancer, while carriers of a single altered copy have increased risk of breast, pancreatic, prostate, and other cancers. Somatic ATM mutations also occur in lymphoid malignancies and solid tumors, and ATM-deficient cancers are often especially sensitive to DNA-damaging treatments.
Consequence/LOF assessment for ATM NM_000051.4:c.7308-9C>T, an intronic substitution 9 nt upstream of the exon 50 acceptor site (11:108330205C>T GRCh38 / 11:108200932C>T GRCh37), predicted protein consequence NP_000042.3:p.?.
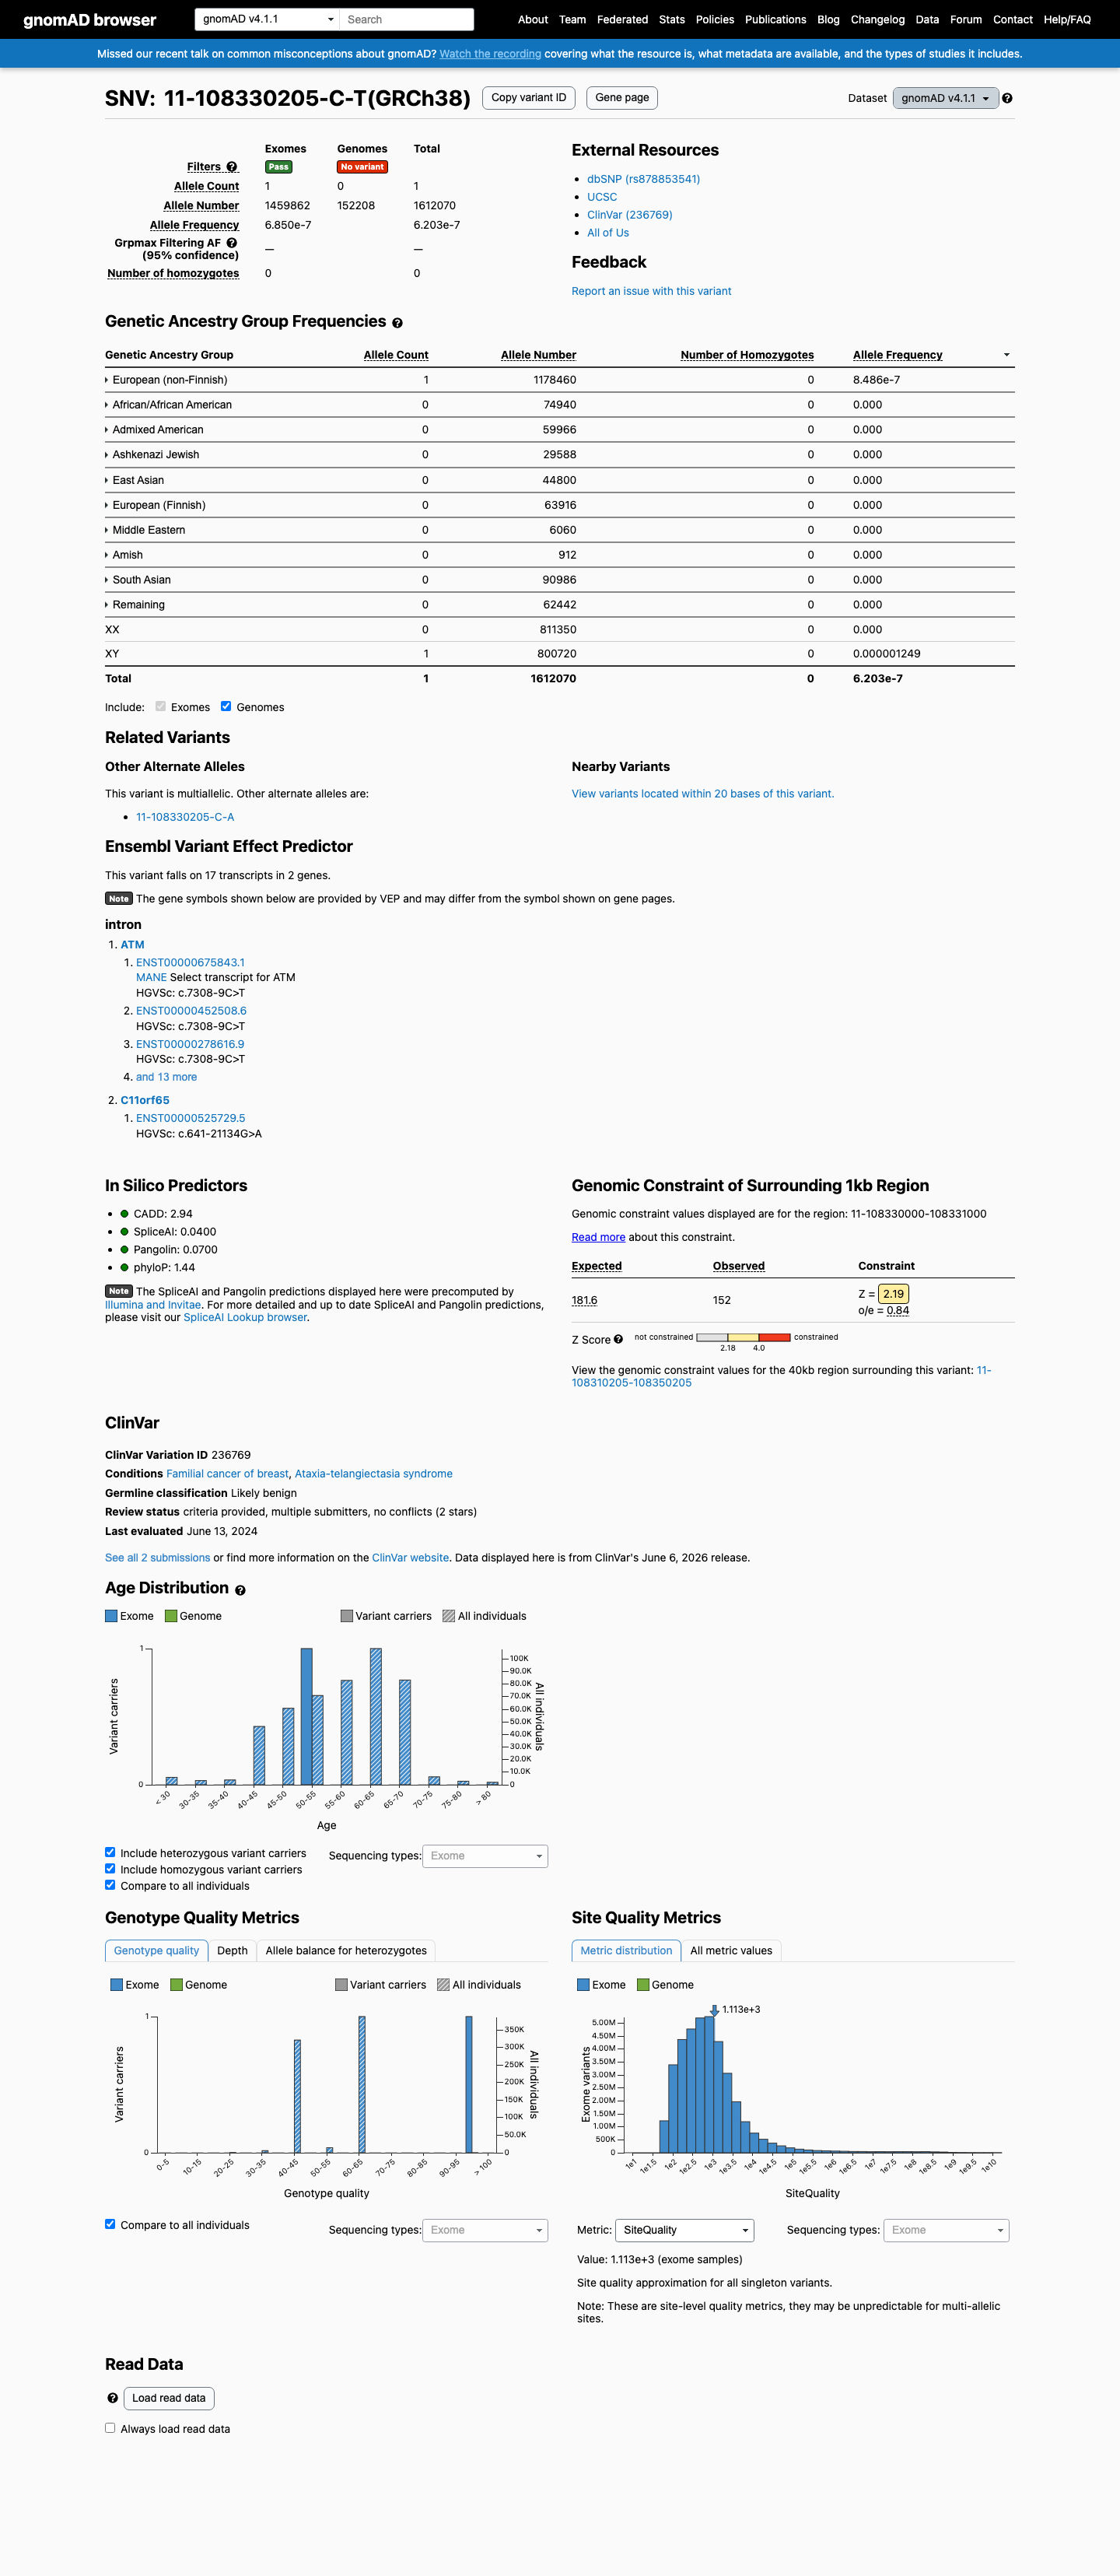
0 hom
European (non-Finnish) 1 / 1,178,460 |
8.5e-05% |
0 hom
0 hom