Back
NM_000059.4:c.6444T>C
p.Ser2148= · BRCA2
0%
complete
Final classification
Likely Benign
BS1BP1BP6
BRCA2
c.6444T>C
p.Ser2148=
synonymous · exon 11

BRCA2 encodes a DNA repair protein that maintains genome stability by repairing double-strand breaks through homologous recombination and by protecting DNA replication forks. It acts as a tumor suppressor, and inherited loss-of-function changes cause hereditary breast and ovarian cancer syndrome, with elevated lifetime risks of breast, ovarian, prostate, and pancreatic cancers; biallelic changes cause Fanconi anemia complementation group D1. Reduced or altered BRCA2 activity is implicated in multiple tumor types, and PARP inhibitors are an approved treatment for BRCA2-associated ovarian and breast cancers.

This variant

As a synonymous BRCA2 change, this variant does not alter the DNA repair protein's amino-acid sequence, while inherited loss of BRCA2 function causes hereditary breast and ovarian cancer predisposition.

Transcript
NM_000059.4
HGVS · transcript:coding
NM_000059.4:c.6444T>C
GRCh38
chr13:32340799 T>C
GRCh37
chr13:32914936 T>C
Likely Benign: BS1 supporting (+1) is outweighed by BP1 strong (-4) and BP6 supporting (-1), yielding ENIGMA's -4 Likely Benign range.
Classification rationale
BS1BP1BP6 Likely Benign
BRCA2 c.6444T>C synonymous · exon 11

Likely Benign: BS1 supporting reflects the maximum non-cancer gnomAD FAF of 0.00007569. Likely Benign: BP1 strong applies to the synonymous change outside approved BRCA2 domains with SpliceAI max delta 0.01. Likely Benign: BP6 supporting is supported by the exact ClinVar expert-panel Likely benign classification.

BS1 + BP1 + BP6 → Likely Benign
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_000059.4 · variants mapped to exon structure
BRCA2 NM_000059.4
Fetching transcript structure from UCSC…
Applied criteria · 3 applied · 13 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 3
Strength Supporting Moderate Strong Very strong
BS1 supporting Benign
Met at supporting: maximum non-cancer gnomAD FAF 0.00007569 falls between the ENIGMA BS1 Supporting limits of greater than 0.00002 and at most 0.0001.
ENIGMA v1.2 specifies BS1 Strong above 0.0001 and BS1 Supporting above 0.00002 through 0.0001, using maximum non-founder gnomAD FAF and separate exome/genome assessment.The non-cancer gnomAD v2.1 exome result is 6/215906 alleles, AF 2.778987151816068e-05, with maximum population FAF 7.568999999999996e-05 and zero homozygotes.The variant is absent from gnomAD v3.1 non-cancer, so the observed v2.1 non-cancer FAF remains the applicable maximum reported FAF.
BP1 strong review Benign
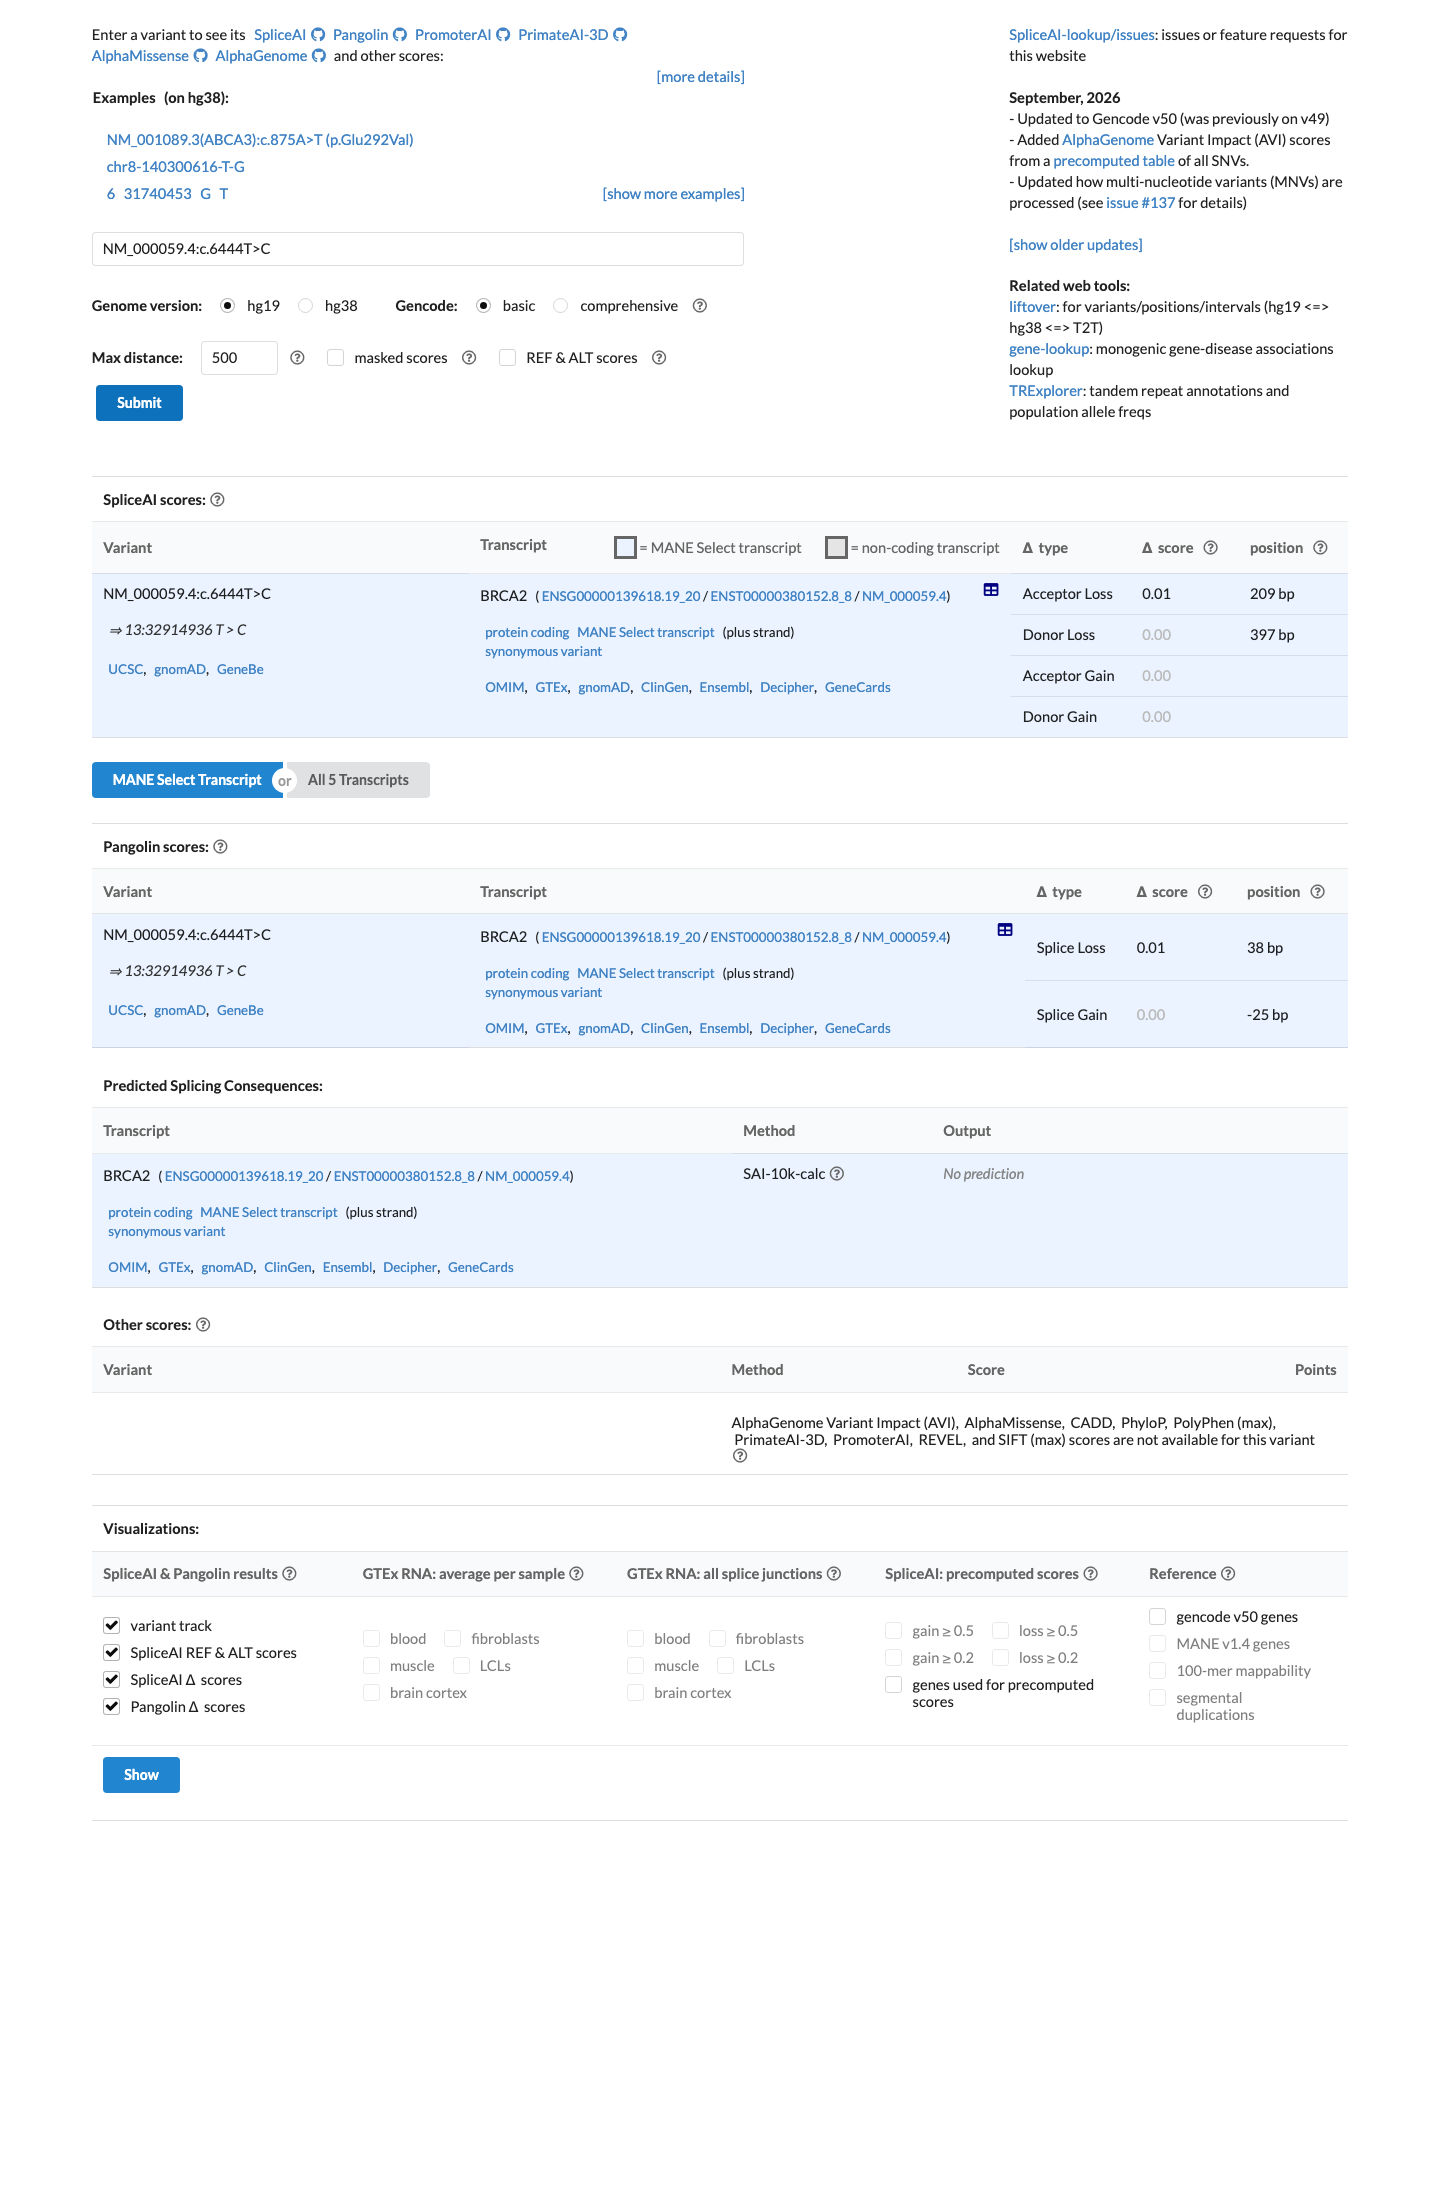
Met at strong: synonymous residue Ser2148 is outside approved domains, and SpliceAI maximum delta 0.01 meets the <=0.1 threshold.
ENIGMA specification v1.2 assigns BP1_Strong to silent substitutions outside a potentially clinically important functional domain with SpliceAI <=0.1.The approved BRCA2 domains are PALB2 binding aa 10-40 and DNA binding aa 2481-3186; residue Ser2148 is outside both domains.SpliceAI reports DS_AG 0.00, DS_AL 0.01, DS_DG 0.00, and DS_DL 0.00; maximum delta score 0.01 is below the <=0.1 threshold.
BP6 supporting Benign
Met, Supporting: ClinVar expert panel classifies the exact variant as Likely benign, satisfying the requested BP6 expert-panel rule.
ClinVar variation 415692 exactly matches NM_000059.4(BRCA2):c.6444T>C (p.Ser2148=) and reports Likely benign with expert-panel review status.The requested BP6 rule allows Supporting BP6 only for an exact-variant ClinVar expert-panel classification of Benign or Likely benign; this condition is satisfied.Six additional laboratory submissions and aggregate ClinVar labels were not used as the trigger.
Assessed · not applied · 4 not met · 9 not assessed
Pathogenic
PS3 Not assessed: no variant-specific damaging functional assay or governing ENIGMA Table 9 entry was found for c.6444T>C (p.Ser2148=).
PS4 Not assessed: no exact-variant case-control p-value, OR, and confidence interval were available to compare with the ENIGMA PS4 thresholds.
PM2 Not met: the variant is present at 6/215906 alleles in non-cancer gnomAD v2.1 exomes, so ENIGMA's required absence from both datasets is not satisfied.
PM3 Not assessed: no documented Fanconi anemia phenotype, chromosome-breakage result, or phased pathogenic/likely pathogenic BRCA2 variant supports the VCEP PM3 requirement.
PP1 Not assessed: no family-level segregation data or quantitative LR was available, so ENIGMA PP1 thresholds of 2.08, 4.3, 18.7, and 350 cannot be applied.
PP3 Not met: SpliceAI max delta 0.01 is below the ENIGMA PP3 threshold of 0.2 for predicted splicing in silent variants.
PP4 Not assessed: no exact-variant multifactorial likelihood ratio was available to compare with the ENIGMA PP4 threshold of >=2.08.
PP5 Not met: the exact ClinVar expert-panel classification is Likely benign, whereas PP5 requires Pathogenic or Likely pathogenic expert-panel classification.
Benign
BA1 Not met: maximum non-cancer gnomAD FAF is 0.00007569, below the ENIGMA BA1 threshold of greater than 0.001.
BS2 Not assessed: gnomAD reports zero homozygotes, but ENIGMA requires phenotyped clinical or research cohort observations rather than population-frequency homozygotes for BS2.
BS3 Not assessed: no variant-specific benign functional assay or governing ENIGMA Table 9 entry was found for c.6444T>C (p.Ser2148=).
BS4 Not assessed: no affected-relative non-segregation data or quantitative LR was available, so ENIGMA BS4 thresholds of 0.48, 0.23, 0.05, and 0.00285 cannot be applied.
BP5 Not assessed: no exact-variant multifactorial LR was available for comparison with the BP5 Supporting threshold of <=0.48.
N/A · 12 PVS1 · PS1 · PS2 · PM1 · PM4 · PM5 · PM6 · PP2 · BP2 · BP3 · BP4 · BP7
Research & evidence
Population frequency · supports benign
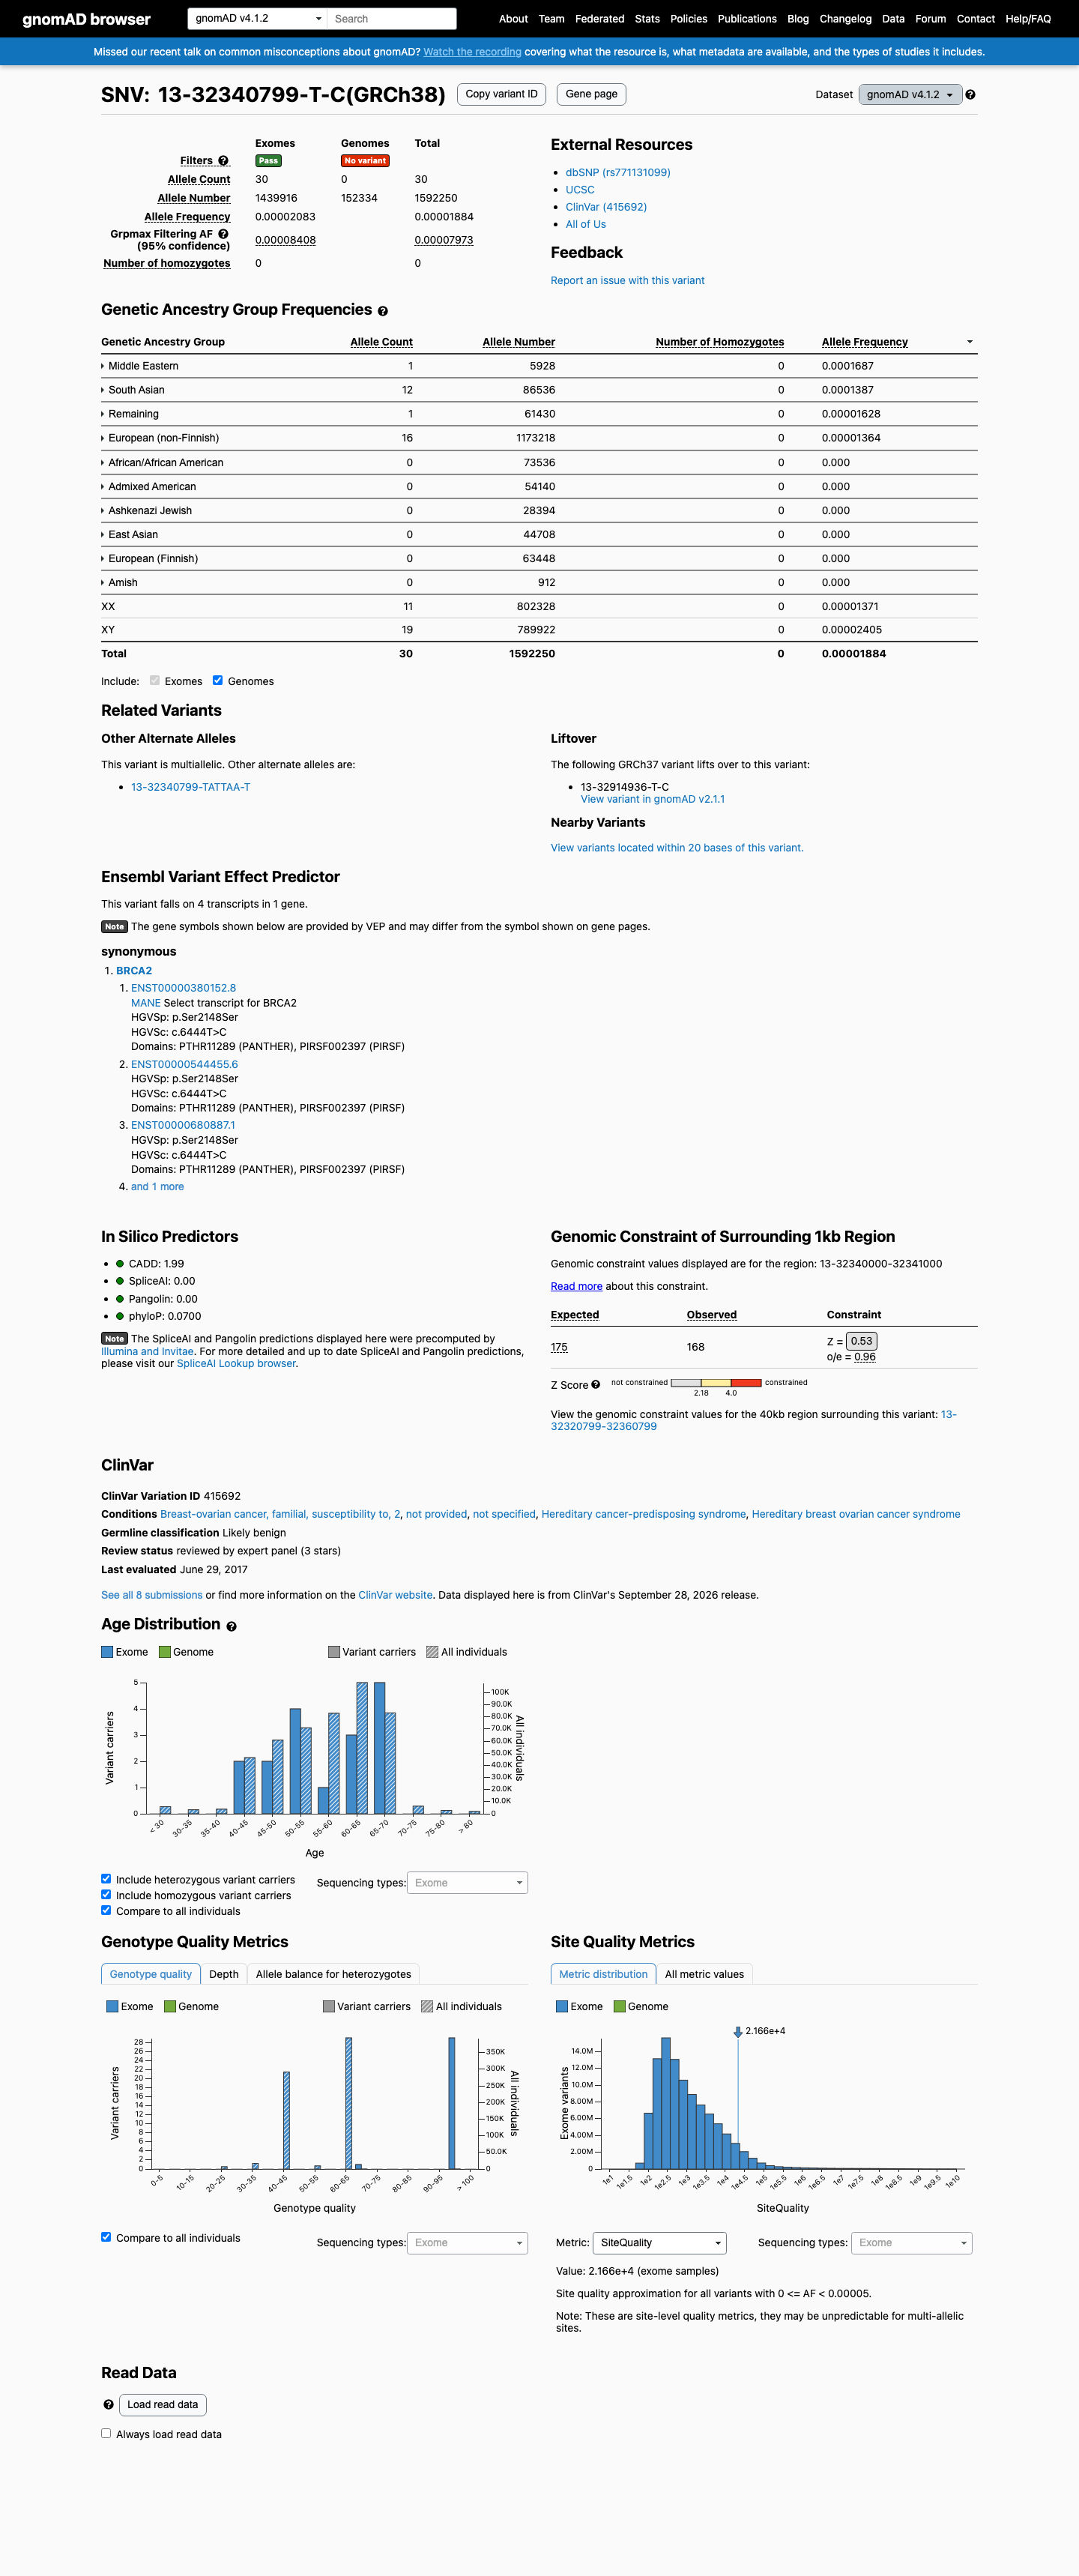
gnomAD v4.1 screenshot
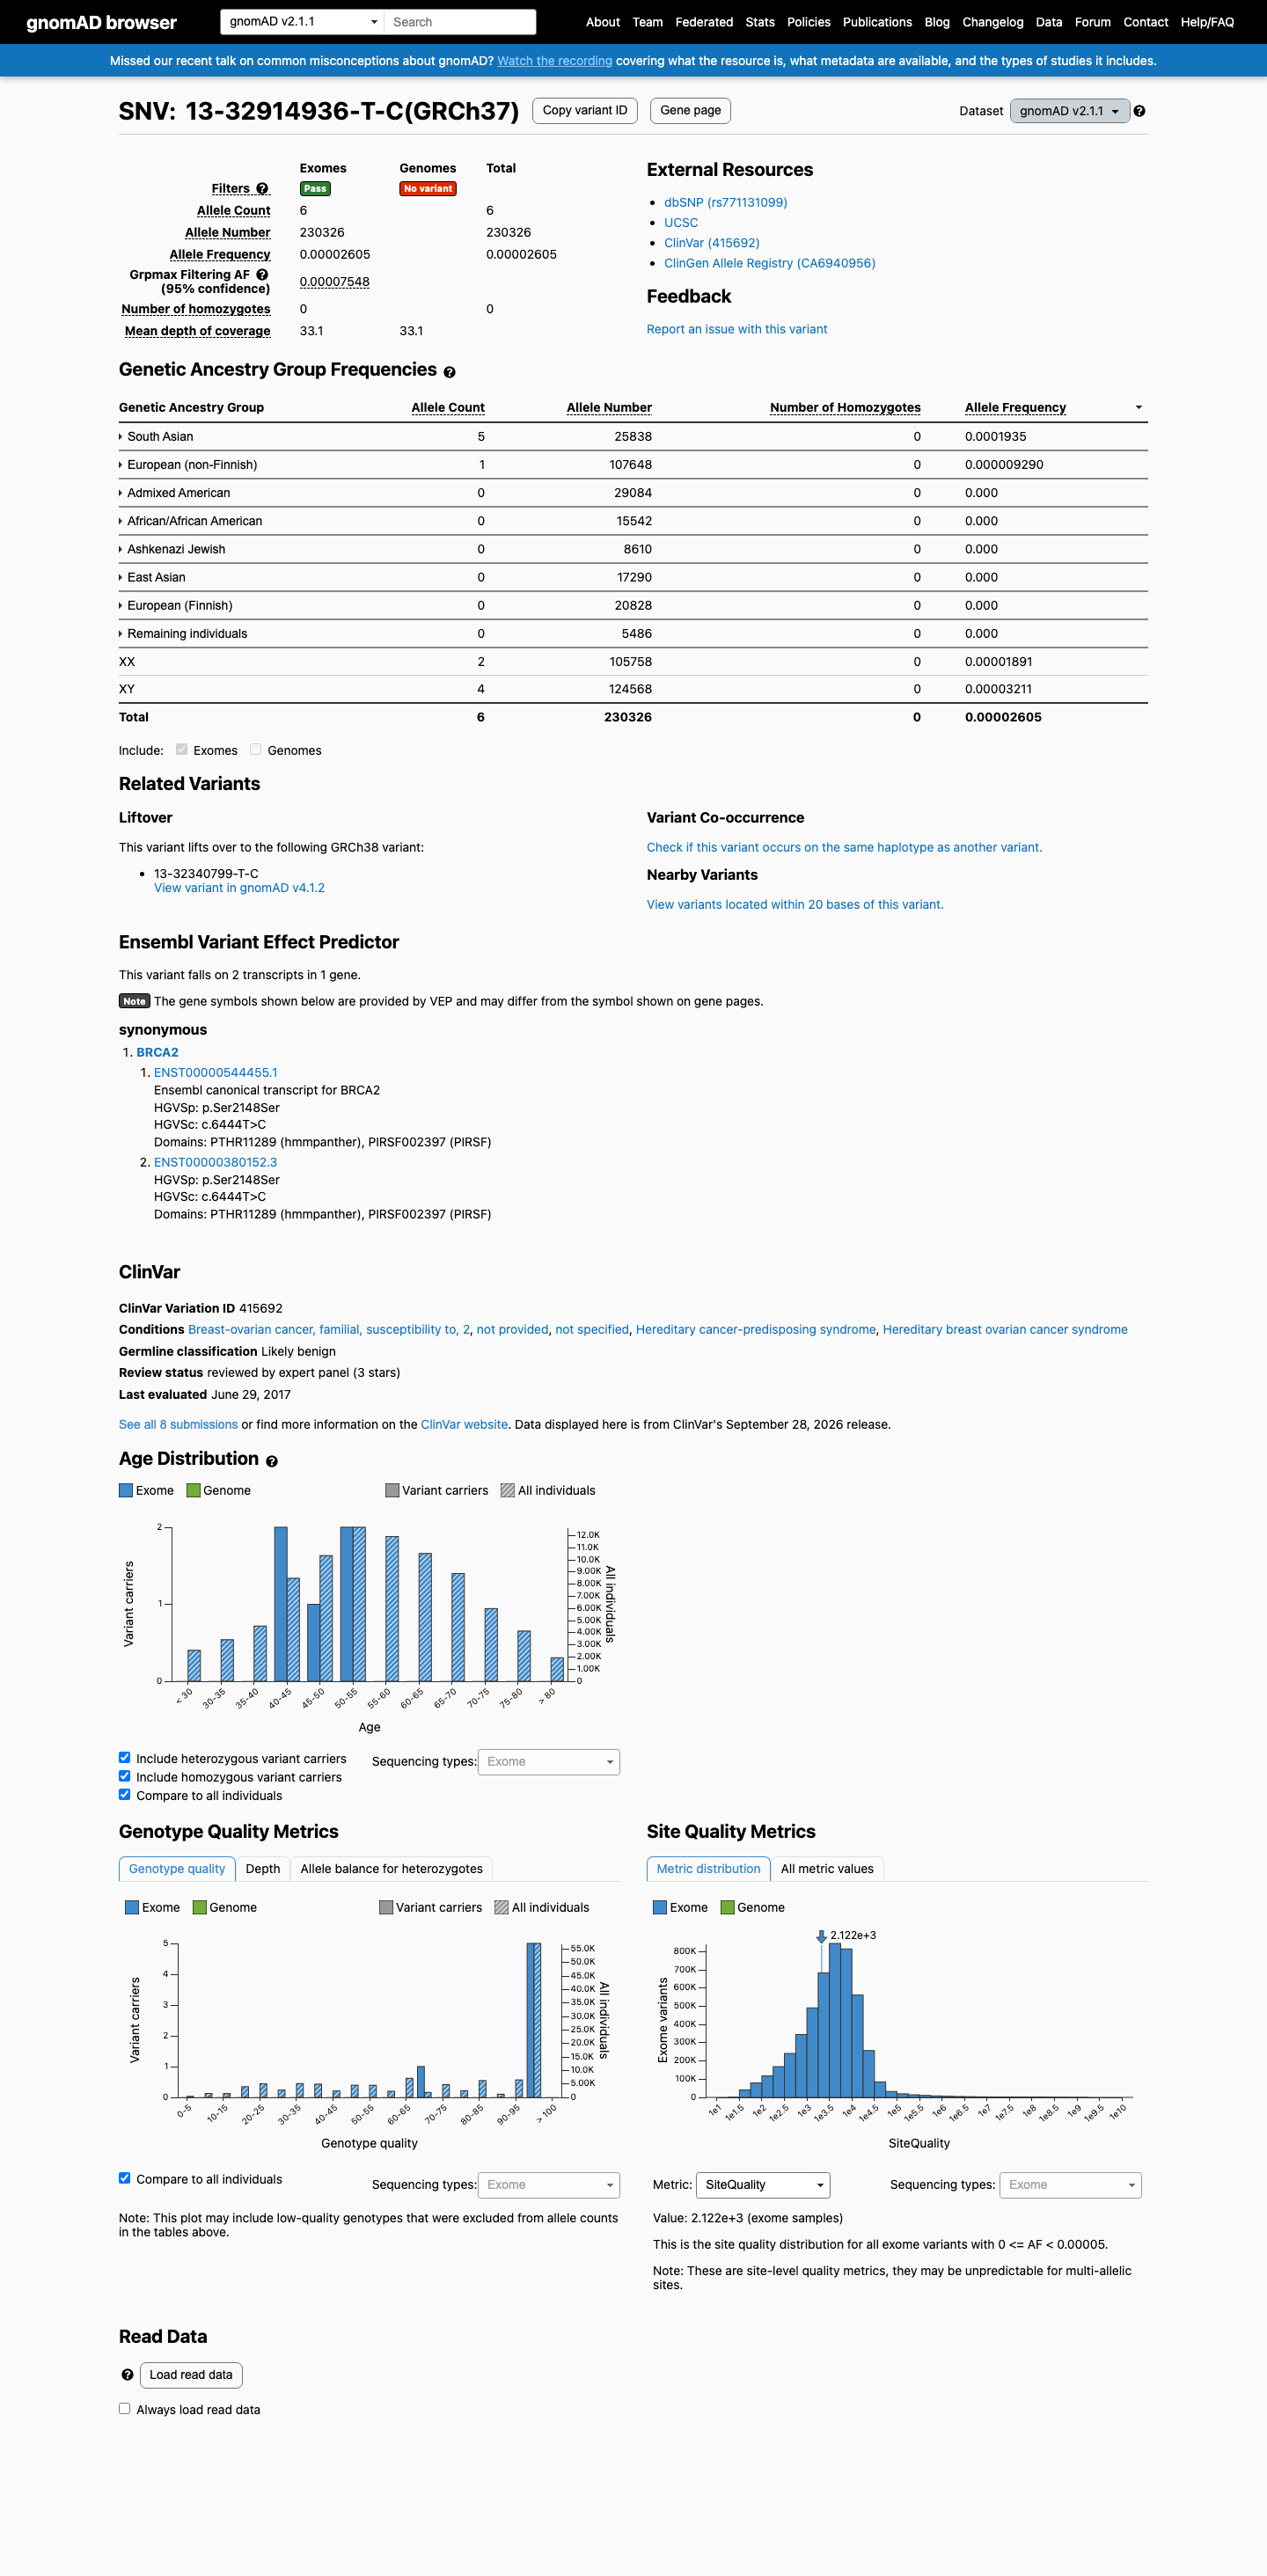
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 1.88413e-05; MAF= 0.00188%, 30/1592250 alleles, homozygotes = 0) and has highest observed frequency in the Middle Eastern population (AF= 0.000168691; MAF= 0.01687%, 1/5928 alleles, homozygotes = 0); grpmax FAF= 7.973e-05.
v2.1
This variant is present in gnomAD v2.1 (AF= 2.605e-05; MAF= 0.00261%, 6/230326 alleles, homozygotes = 0) and has highest observed frequency in the South Asian population (AF= 0.000193513; MAF= 0.01935%, 5/25838 alleles, homozygotes = 0); grpmax FAF= 7.548e-05.
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.0019% · 30 / 1,592,250
0 hom · FAF 0.008%
Middle Eastern
1 / 5,928
0.017%
South Asian
12 / 86,536
0.014%
Remaining individuals
1 / 61,430
0.0016%
European (non-Finnish)
16 / 1,173,218
0.0014%
+ 6 not observed (Admixed American, European (Finnish), Amish, East Asian, Ashkenazi Jewish, African/African American)
gnomAD v2.1
0.0026% · 6 / 230,326
0 hom · FAF 0.0075%
South Asian
5 / 25,838
0.019%
European (non-Finnish)
1 / 107,648
0.00093%
+ 6 not observed (African/African American, Admixed American, Ashkenazi Jewish, East Asian, European (Finnish), Remaining individuals)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
ClinVar screenshot
ClinVar
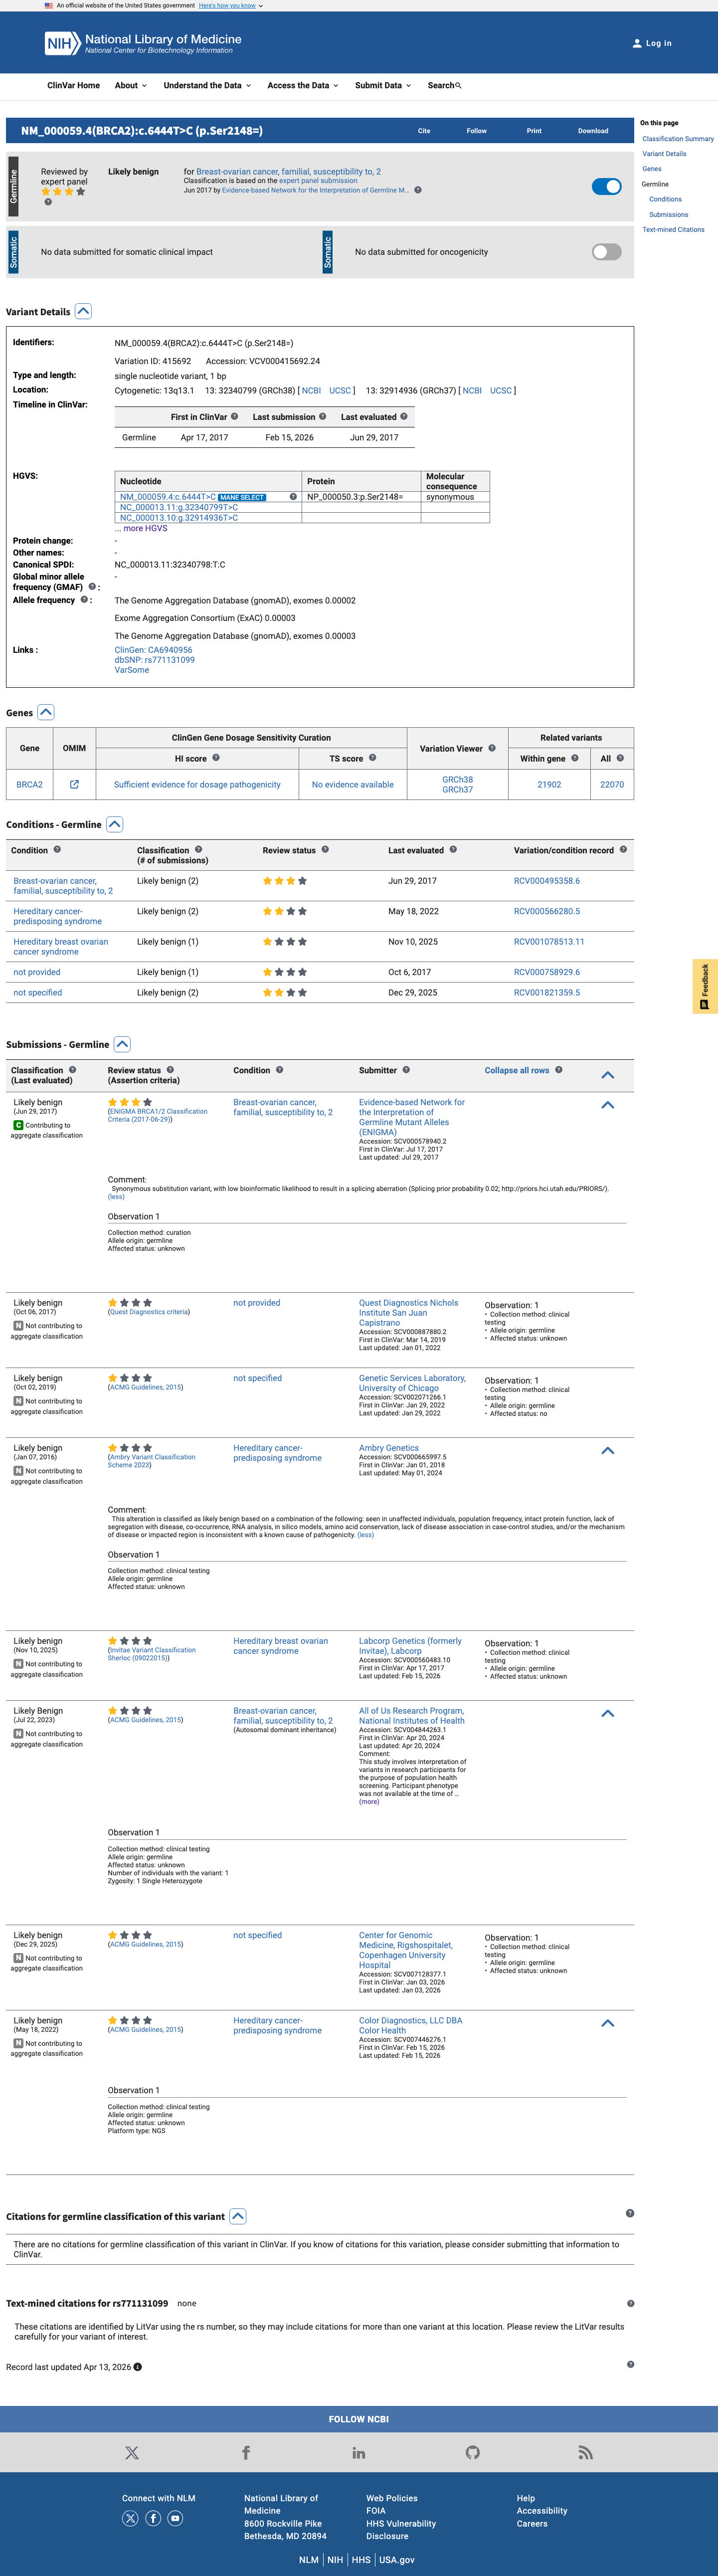
This variant has been reported in ClinVar as Likely benign (6 clinical laboratories) and as Likely Benign (1 clinical laboratory) and as Likely benign by Evidence-based Network for the Interpretation of Germline Mutant Alleles (ENIGMA) (expert panel). (ClinVarID = 415692)
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.01).
Functional / OncoKB screenshot
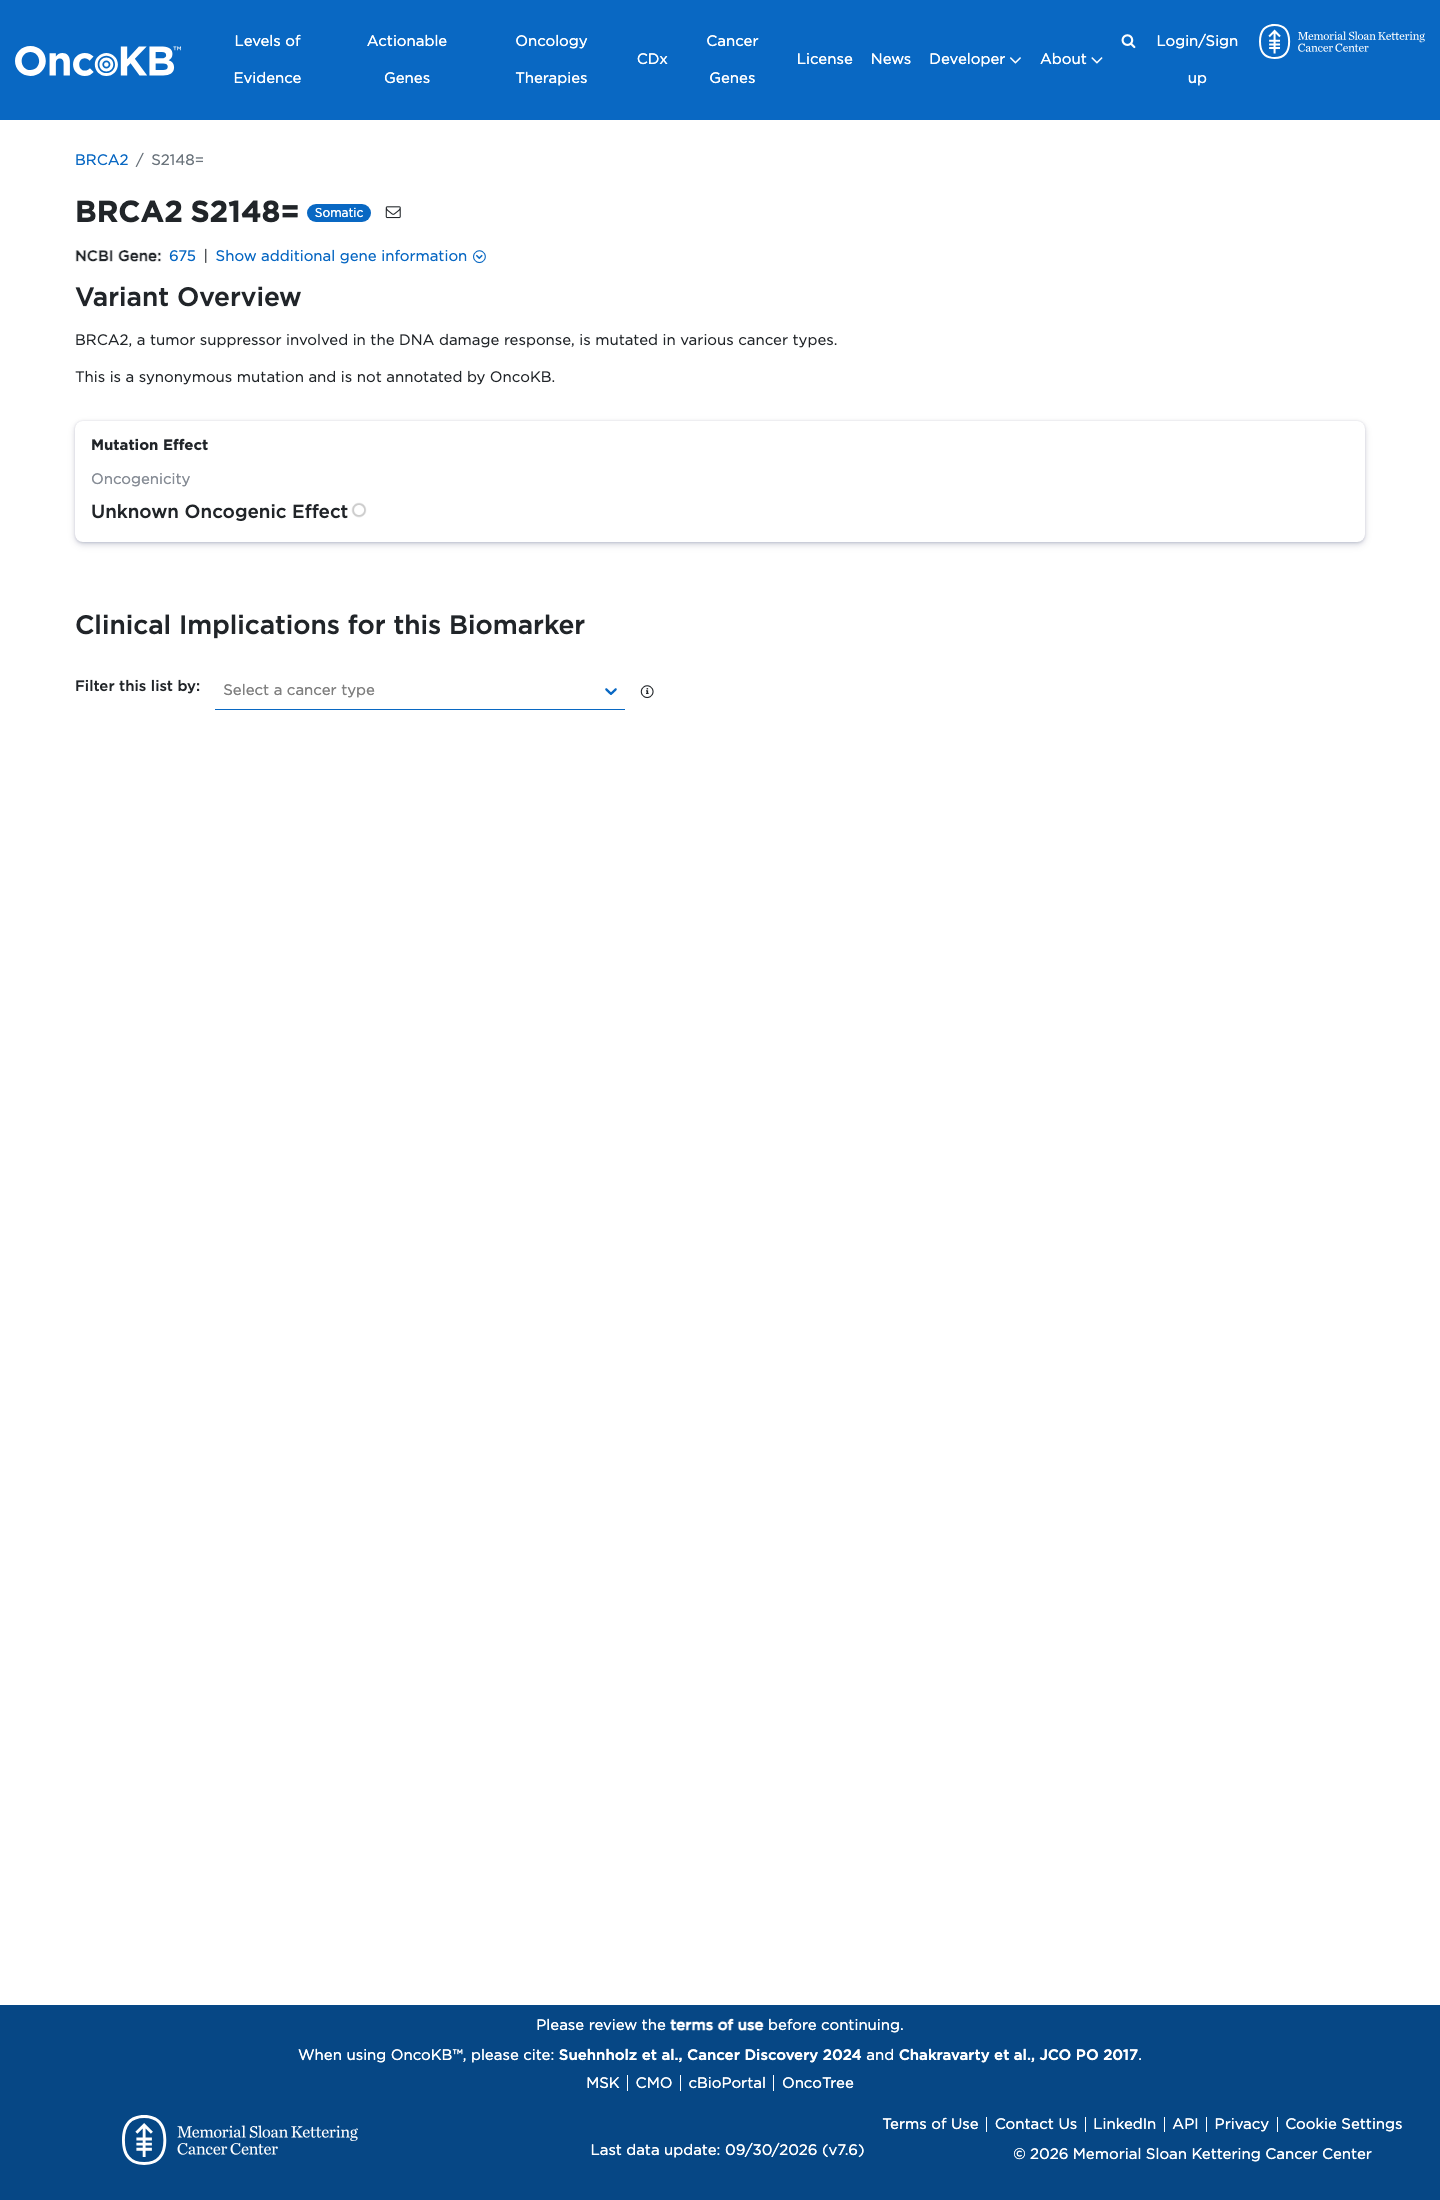
Functional Unknown Oncogenic Effect
OncoKB identified curated literature and non-variant-specific oncogenicity context for review; listed oncogenicity label: Unknown Oncogenic Effect.
OncoKB ↗
COSMIC screenshot
COSMIC
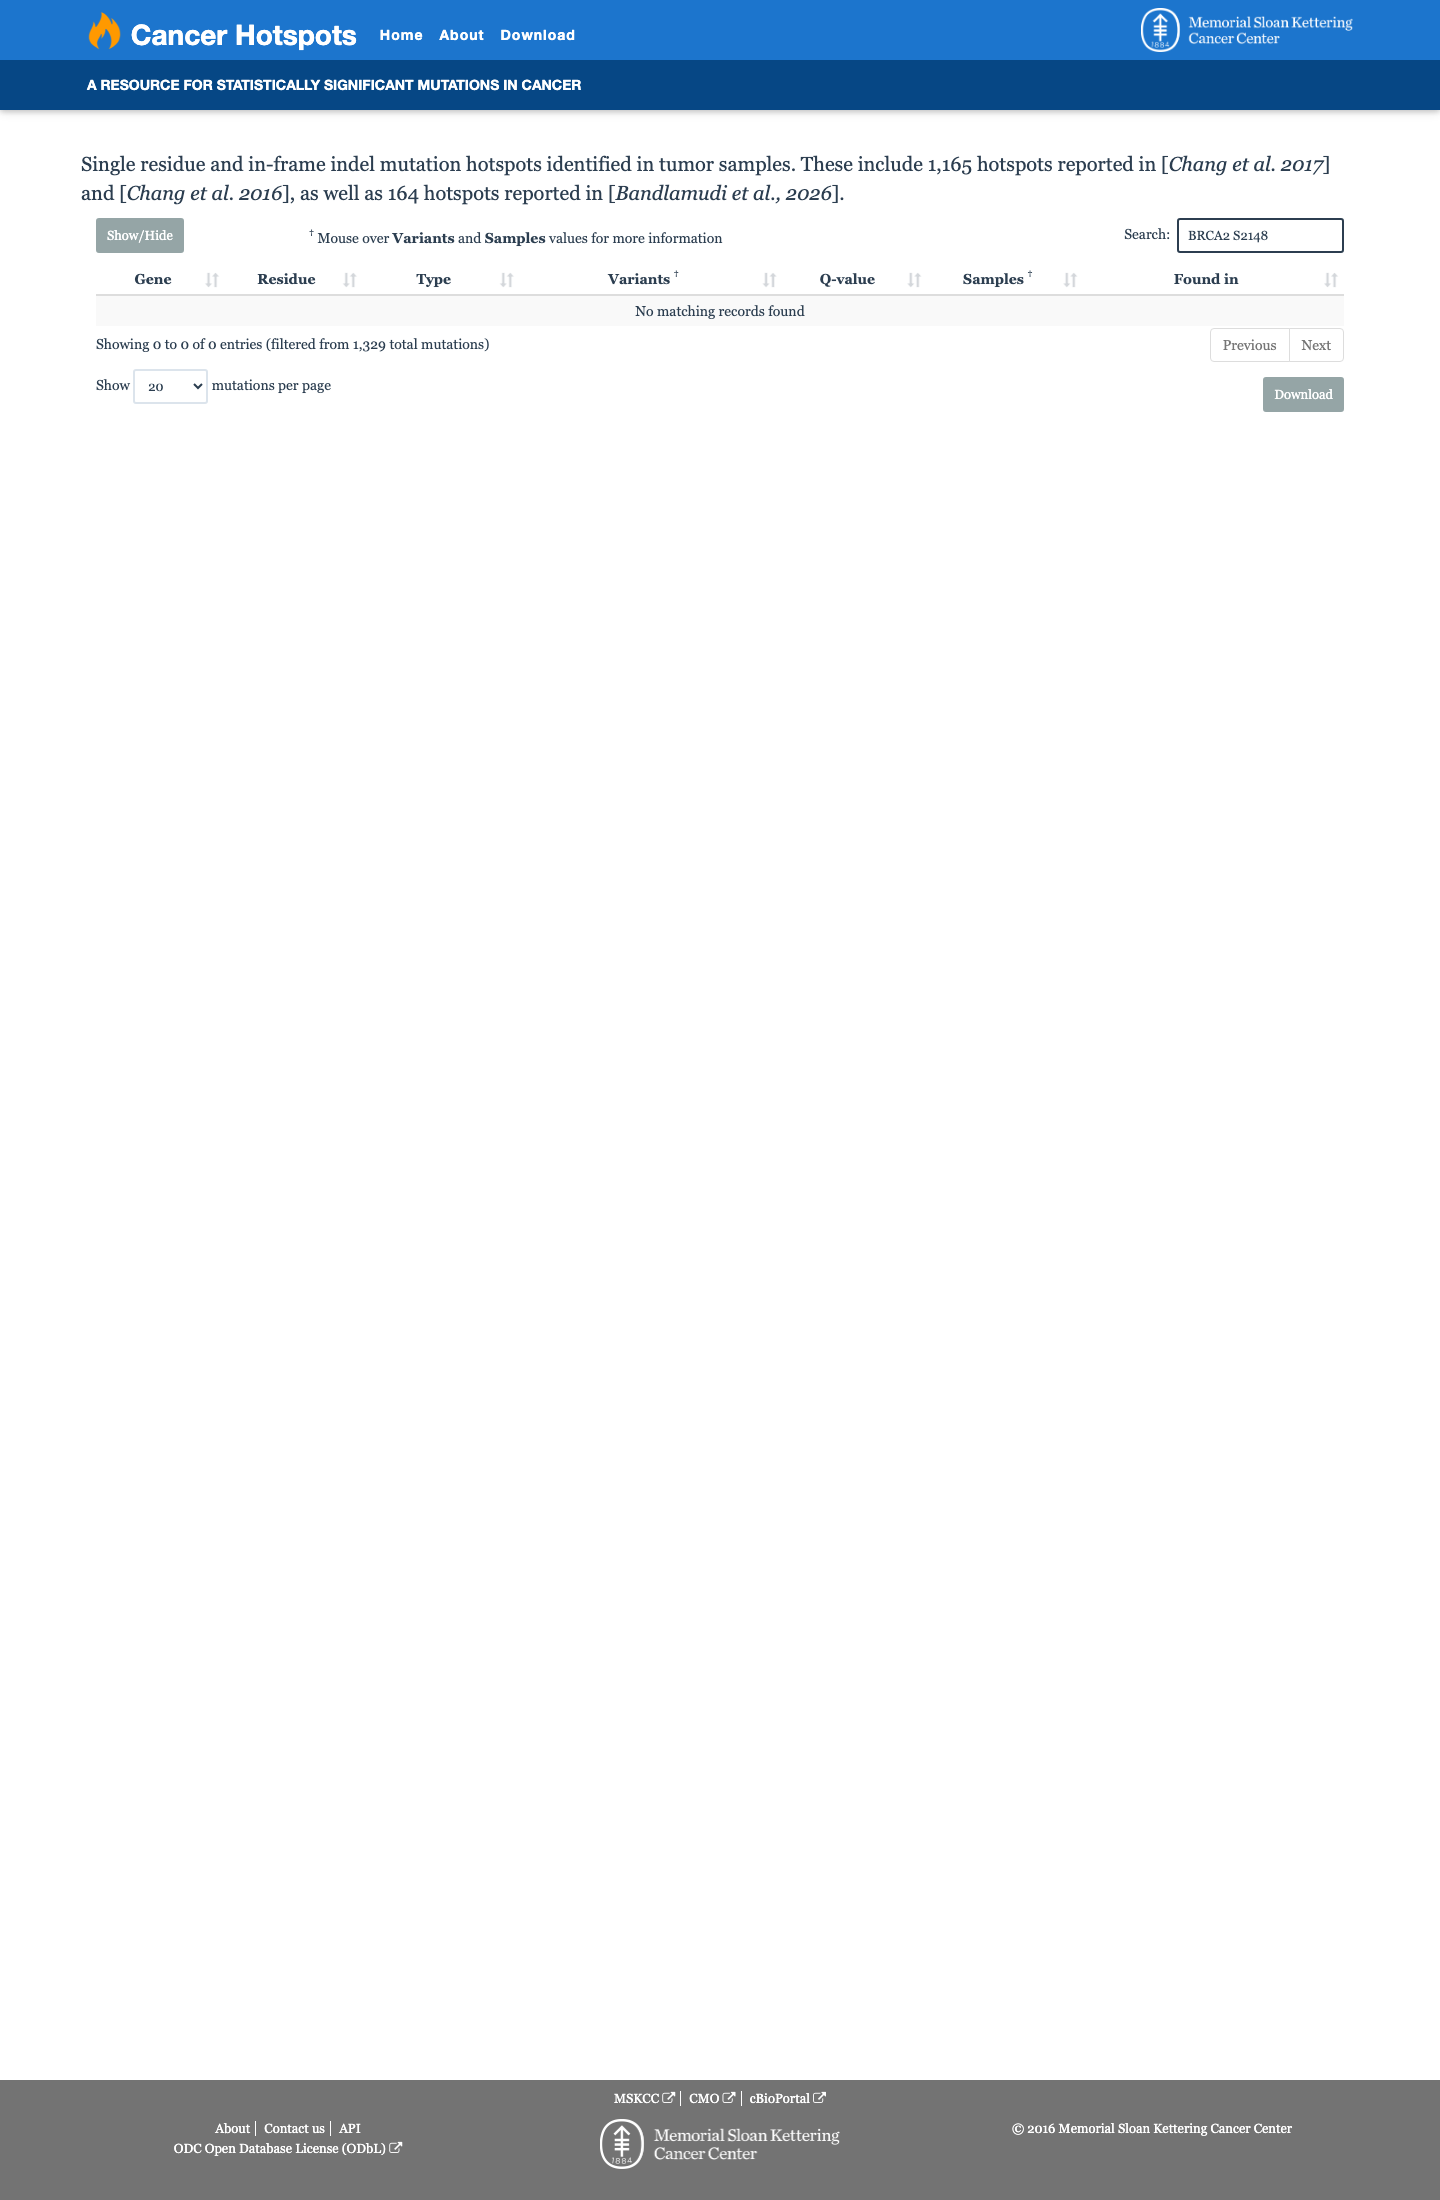
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Sources & reference links
9Sources
CSpec VCEP
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 8 PMIDs not cited in assessment
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology. CLINVAR
26467025 ↗ A Standardized DNA Variant Scoring System for Pathogenicity Assessments in Mendelian Disorders. CLINVAR
18163131 ↗ The emerging landscape of breast cancer susceptibility. CLINVAR
23188549 ↗ NSGC practice guideline: risk assessment and genetic counseling for hereditary breast and ovarian cancer. CLINVAR
25356965 ↗ ACMG policy statement: updated recommendations regarding analysis and reporting of secondary findings in clinical genome-scale sequencing. CLINVAR
27854360 ↗ Recommendations for reporting of secondary findings in clinical exome and genome sequencing, 2016 update (ACMG SF v2.0): a policy statement of the American College of Medical Genetics and Genomics. CLINVAR
34012068 ↗ ACMG SF v3.0 list for reporting of secondary findings in clinical exome and genome sequencing: a policy statement of the American College of Medical Genetics and Genomics (ACMG). CLINVAR
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification criteria. CLINVAR