Back
NM_000059.4:c.9738C>T
p.Ala3246= · BRCA2
ClinGen ENIGMA BRCA1 and BRCA2 Expert Panel Specifications to the ACMG/AMP Variant Interpretation Guidelines for BRCA2 Version 1.2 · v1.2
0%
complete
Final classification
Benign
BA1BS1BP1BP4BP6BP7
BRCA2
c.9738C>T
p.Ala3246=
synonymous · exon 27

BRCA2 encodes a DNA repair protein that maintains genome stability by repairing double-strand breaks through homologous recombination and by protecting DNA replication forks. It acts as a tumor suppressor, and inherited loss-of-function changes cause hereditary breast and ovarian cancer syndrome, with elevated lifetime risks of breast, ovarian, prostate, and pancreatic cancers; biallelic changes cause Fanconi anemia complementation group D1. Reduced or altered BRCA2 activity is implicated in multiple tumor types, and PARP inhibitors are an approved treatment for BRCA2-associated ovarian and breast cancers.

This variant

BRCA2 encodes a tumor-suppressor DNA-repair protein involved in homologous recombination and replication-fork protection, and inherited loss of function causes hereditary breast and ovarian cancer syndrome.

Transcript
NM_000059.4
HGVS · transcript:coding
NM_000059.4:c.9738C>T
GRCh38
chr13:32398251 C>T
GRCh37
chr13:32972388 C>T
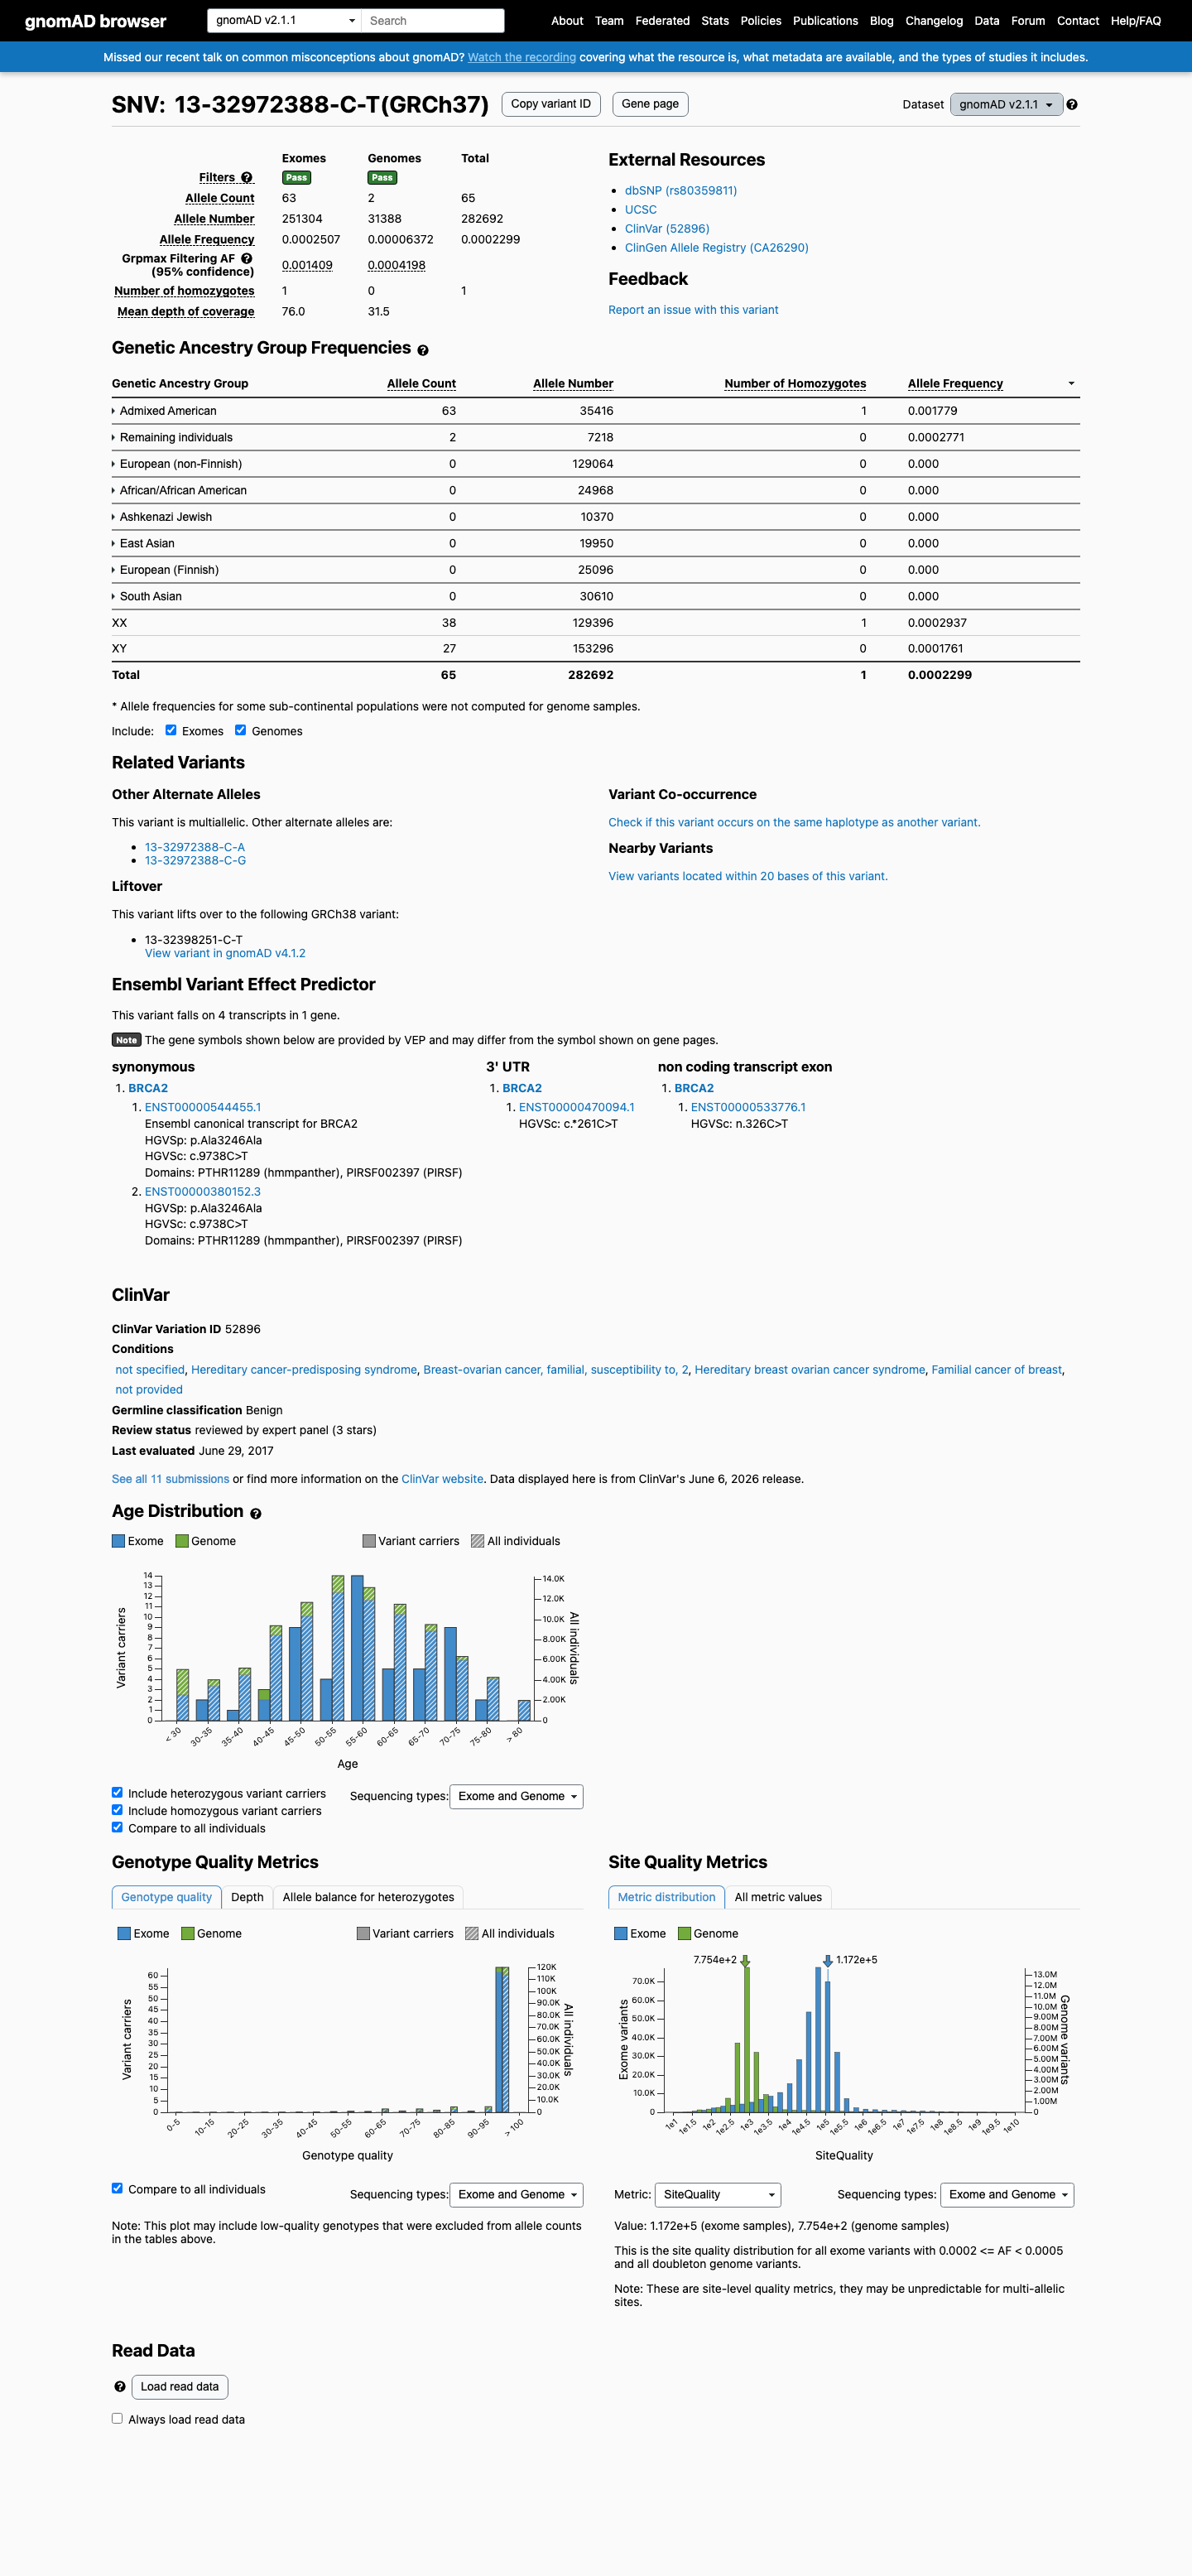
Benign: BA1 stand-alone benign is met by the gnomAD v2.1 non-cancer exome maximum filter allele frequency of 0.00142293, exceeding the ENIGMA threshold of 0.001.
Classification rationale
BA1BS1BP1BP4BP6BP7 Benign
BRCA2 c.9738C>T synonymous · exon 27

Benign: BA1 is met by the gnomAD v2.1 non-cancer exome maximum filter allele frequency of 0.00142293, above 0.001. Benign: BS1 strong is met because the maximum filter allele frequency of 0.00142293 exceeds 0.0001. Benign: BP1 strong is met for the synonymous Ala3246 change outside the clinically important domains without predicted splice impact. Benign: BP4 supporting is met because SpliceAI maximum delta is 0.013, below 0.1. Benign: BP6 supporting is met by the exact-variant ClinVar expert-panel Benign classification. Benign: BP7 supporting is met because the synonymous variant satisfies the BP4 no-splice-impact prerequisite.

BA1 + BS1 + BP1 + BP4 + BP6 + BP7 → Benign
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_000059.4 · variants mapped to exon structure
BRCA2 NM_000059.4
Fetching transcript structure from UCSC…
Applied criteria · 6 applied · 12 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 6
Strength Supporting Moderate Strong Very strong
BA1 stand-alone Benign
Met: gnomAD v2.1 non-cancer exome maximum filter allele frequency 0.00142293 exceeds the BRCA2 VCEP BA1 threshold of 0.001.
The BRCA2 VCEP specification defines BA1 as maximum filter allele frequency above 0.001 in a gnomAD non-founder population, using gnomAD v2.1 non-cancer exomes and gnomAD v3.1 non-cancer genomes with exome and genome data considered separately.The BRCA2 VCEP appendix states that BA1 was rounded to 0.001 for BRCA2 based on a minimal credible allele frequency of 0.000906, and requires filter allele frequency rather than minor allele frequency.The case evidence reports gnomAD v2.1 non-cancer exome AF 0.0002660855 from 63/236766 alleles, one homozygote, and maximum filter allele frequency 0.00142293; this exceeds 0.001.
BS1 strong Benign
Met, strong: gnomAD v2.1 non-cancer exome maximum filter allele frequency 0.00142293 exceeds the BRCA2 VCEP BS1 threshold of 0.0001.
The BRCA2 VCEP specification defines BS1 as maximum filter allele frequency above 0.0001 and BS1_Supporting as greater than 0.00002 through 0.0001 in a gnomAD non-founder population, with exome and genome data assessed separately.The BRCA2 VCEP appendix derives 0.0001 as the rounded BS1 threshold for BRCA2 and recommends the highest filter allele frequency in a non-founder population, excluding TCGA data and established pathogenic founder variants.The case evidence reports gnomAD v2.1 non-cancer exome maximum filter allele frequency 0.00142293 for this variant, exceeding the strong BS1 threshold of 0.0001.
BP1 strong review Benign
Met, Strong: synonymous Ala3246 is outside the aa 2481-3186 DNA-binding domain and SpliceAI max delta 0.013 is below the <=0.1 threshold.
ENIGMA BRCA2 v1.2 states: apply BP1_Strong for a silent substitution outside the potentially clinically important domains when no splicing is predicted (SpliceAI <=0.1).The variant is p.(Ala3246=); Ala3246 is outside the BRCA2 PALB2-binding domain aa 10-40 and DNA-binding domain aa 2481-3186.SpliceAI max delta is 0.013, which is <=0.1 and therefore satisfies the ENIGMA no-predicted-splicing condition.
BP4 supporting Benign
Met, Supporting: SpliceAI maximum delta 0.013 meets the ENIGMA BP4 threshold of <=0.1 for no predicted splice impact.
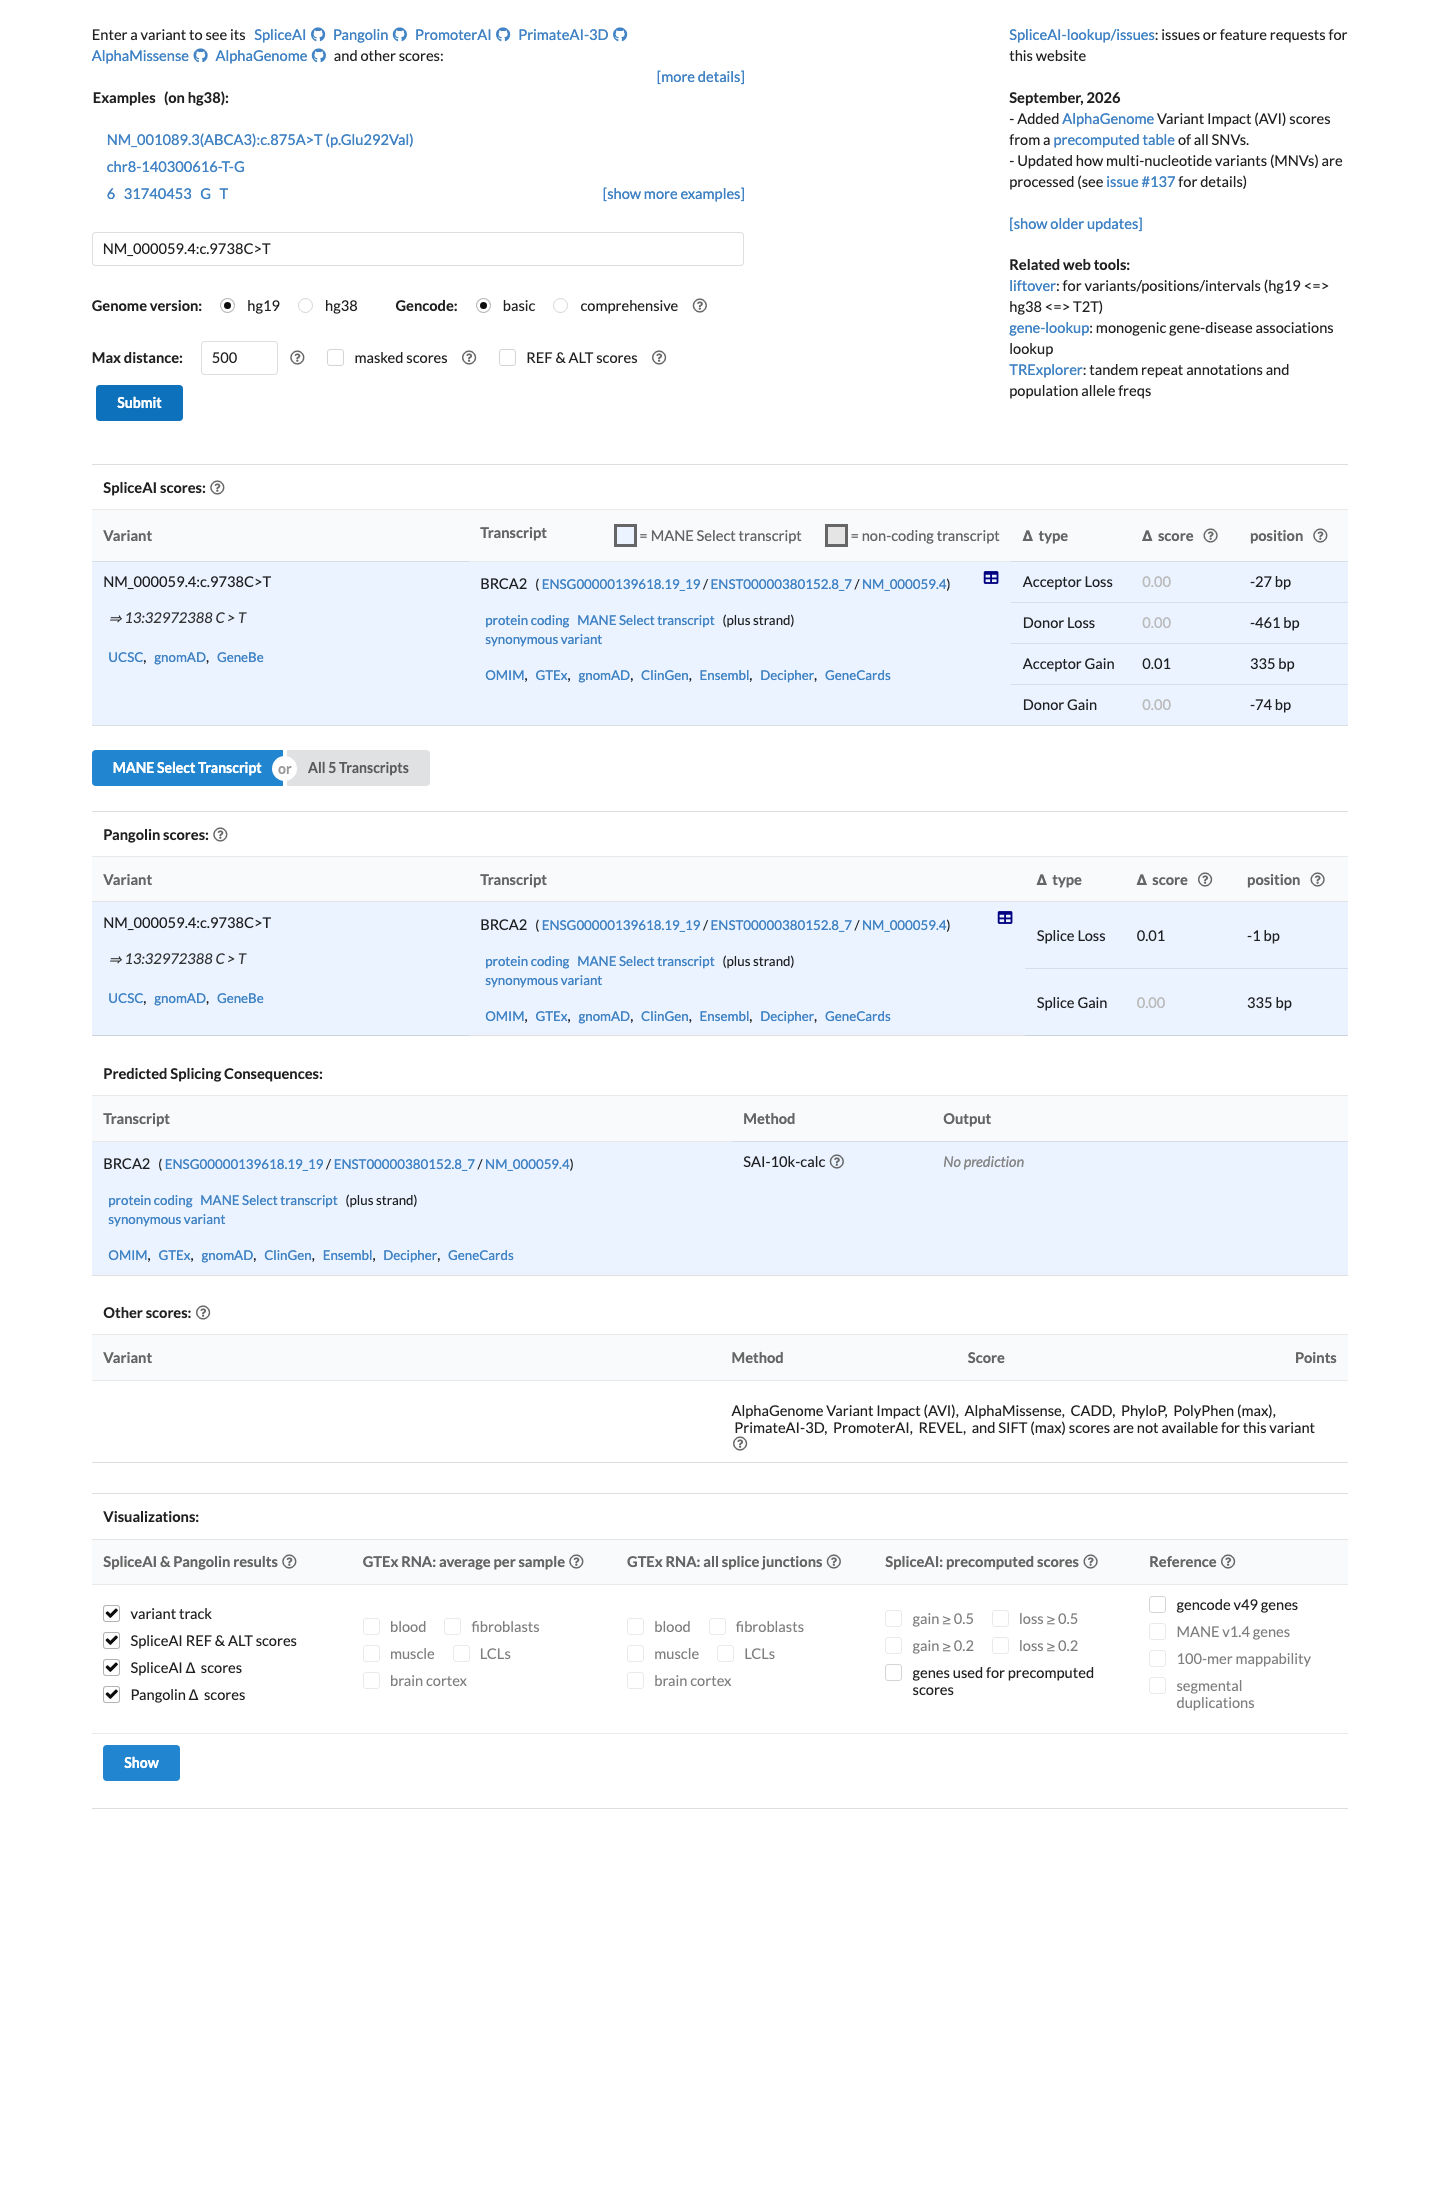
The case normalization identifies NM_000059.4:c.9738C>T as the synonymous protein change NP_000050.3:p.(Ala3246=), so the splice-impact path is applicable and REVEL is not used.The ENIGMA BRCA2 specification states that BP4 applies to a silent variant inside a potentially clinically important functional domain when SpliceAI <=0.1.The case SpliceAI result reports DS_AG 0.013, DS_AL 0.004, DS_DG 0.0, and DS_DL 0.0, with maximum delta 0.013; this meets the governing <=0.1 threshold.
BP6 supporting Benign
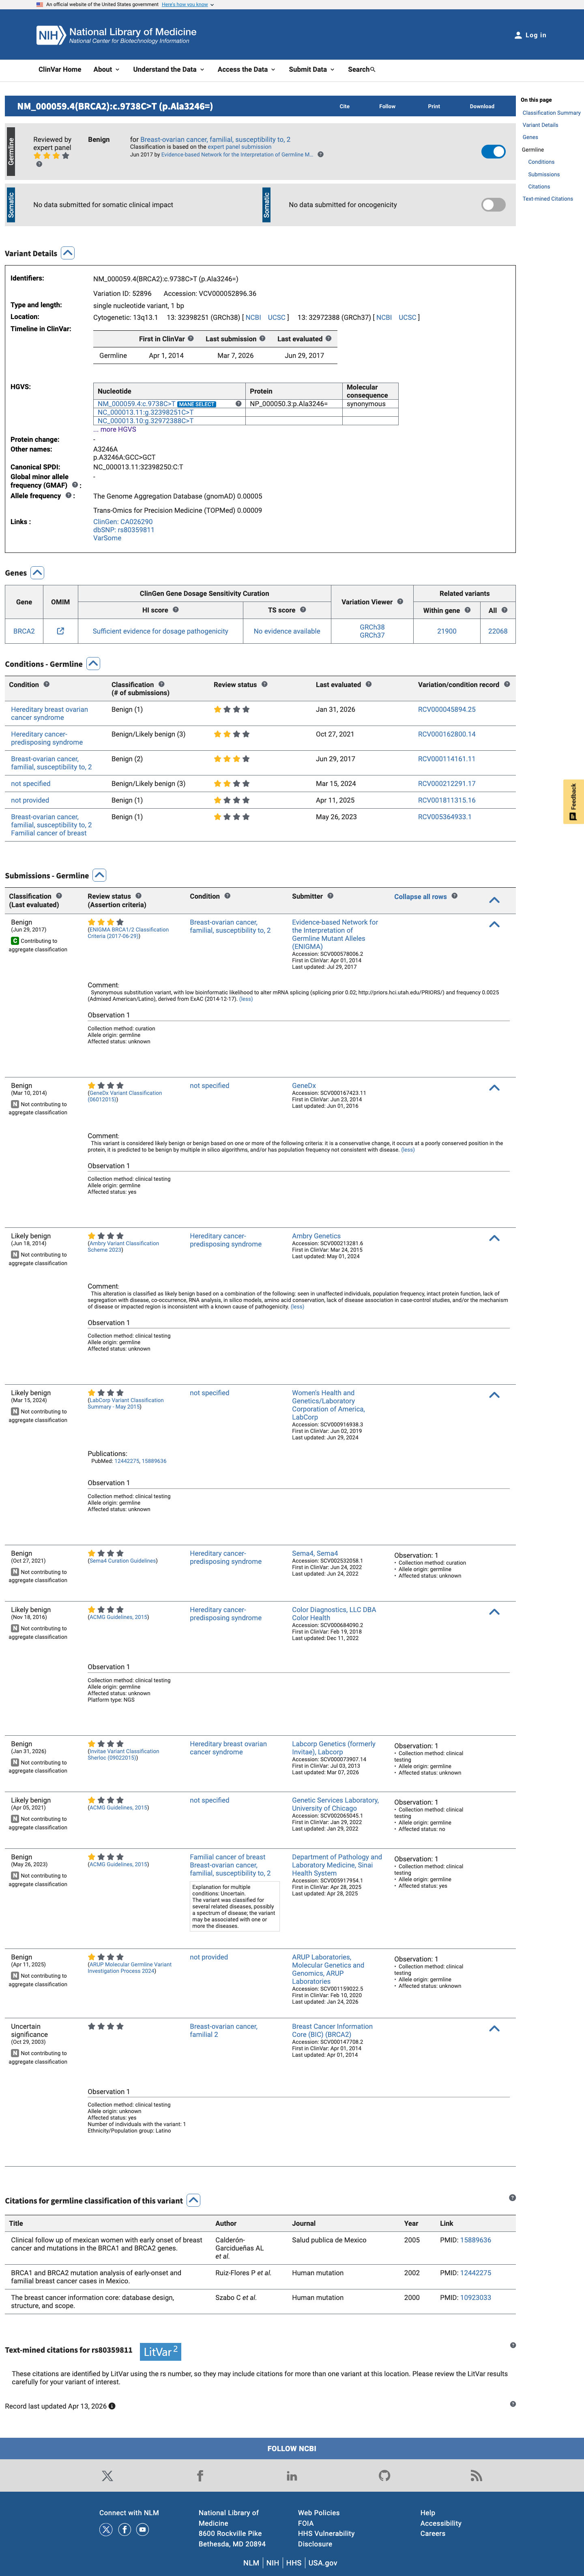
Met, Supporting: exact-variant ClinVar expert-panel classification is Benign for c.9738C>T (p.Ala3246=), satisfying the requested BP6 rule.
ClinVar variation 52896 exactly matches NM_000059.4(BRCA2):c.9738C>T (p.Ala3246=).The ClinVar record identifies the Evidence-based Network for the Interpretation of Germline Mutant Alleles (ENIGMA) as the expert panel and reports Benign with reviewed-by-expert-panel status.The requested exact-variant rule is satisfied: Benign expert-panel classification -> BP6 Supporting.
BP7 supporting Benign
Met, Supporting: synonymous p.Ala3246= has SpliceAI maximum delta 0.013, satisfying BP4 and the ENIGMA BP7 requirement for silent variants.
The case normalization identifies NM_000059.4:c.9738C>T as the synonymous protein change NP_000050.3:p.(Ala3246=).The ENIGMA BRCA2 specification states that BP7 Supporting applies to silent variants inside a potentially clinically important functional domain if BP4 is met, and is applied in addition to BP4.The case SpliceAI maximum delta is 0.013, which meets the ENIGMA BP4 no-impact threshold of <=0.1 and therefore satisfies the BP7 prerequisite.
Assessed · not applied · 4 not met · 8 not assessed
Pathogenic
PS3 Not assessed: no calibrated protein-function or combined mRNA-and-protein assay entry was found for c.9738C>T (p.Ala3246=).
PS4 Not met: published observations lack the ENIGMA PS4 requirements of p-value <=0.05, OR >=4, and a confidence interval excluding 2.0.
PM2 Not met: the variant is present in gnomAD v2.1 non-cancer exomes at AF 0.0002660855 and gnomAD v3.1 non-cancer genomes at AF 0.0000473491.
PM3 Not assessed: no documented Fanconi anemia phenotype, pathogenic BRCA2 co-occurring variant, phase, or PM3-scoring affected-proband observation is available.
PP1 Not assessed: no pedigree or quantitative co-segregation likelihood ratio is available to compare with the ENIGMA PP1 thresholds of 2.08, 4.3, 18.7, or 350.
PP3 Not met: SpliceAI maximum delta 0.013 is below the ENIGMA PP3 threshold of >=0.2 for predicted splicing in silent variants.
PP4 Not assessed: no exact-variant combined clinical likelihood ratio was available for comparison with the PP4 threshold of LR >=2.08.
PP5 Not met: exact-variant ClinVar expert-panel classification is Benign, not Pathogenic or Likely pathogenic, so PP5 is not triggered.
Benign
BS2 Not assessed: one gnomAD homozygote is frequency evidence, not the VCEP's required phenotyped healthy-adult observation for BS2.
BS3 Not assessed: no calibrated benign protein-function or combined mRNA-and-protein assay entry was found for c.9738C>T (p.Ala3246=).
BS4 Not assessed: no affected-relative non-segregation result or quantitative likelihood ratio is available to compare with the ENIGMA BS4 thresholds of 0.48, 0.23, 0.05, or 0.00285.
BP5 Not assessed: no exact-variant combined LR was available for comparison with the BP5 Supporting threshold of LR <=0.48.
N/A · 10 PVS1 · PS1 · PS2 · PM1 · PM4 · PM5 · PM6 · PP2 · BP2 · BP3
Research & evidence
Population frequency · supports benign
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
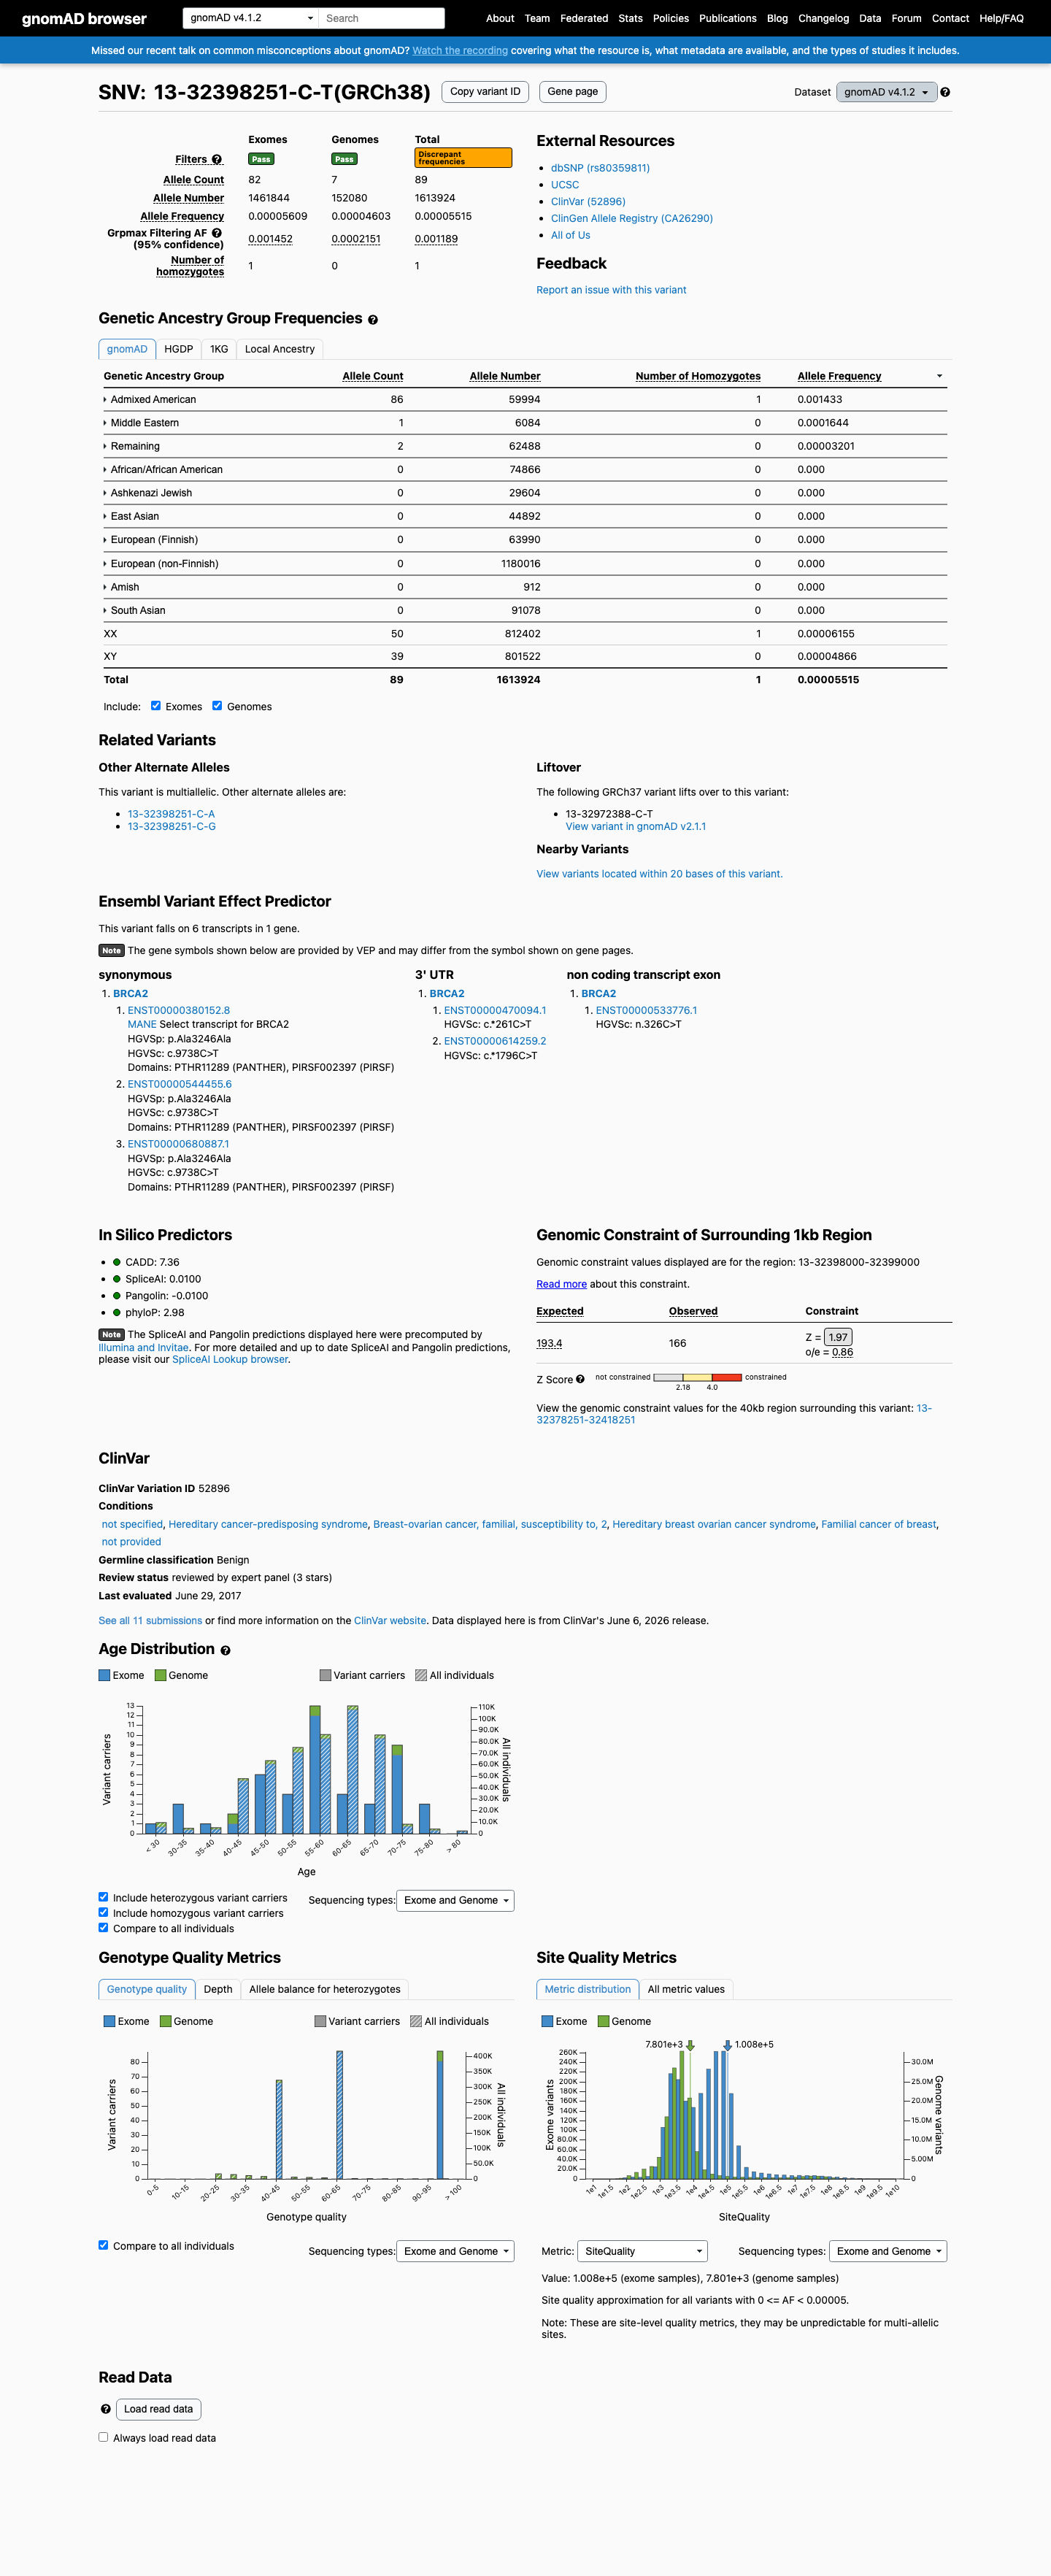
This variant is present in gnomAD v4.1 (AF= 5.51451e-05; MAF= 0.00551%, 89/1613924 alleles, homozygotes = 1) and has highest observed frequency in the Admixed American population (AF= 0.00143348; MAF= 0.14335%, 86/59994 alleles, homozygotes = 1); grpmax FAF= 0.00118865.
v2.1
This variant is present in gnomAD v2.1 (AF= 0.000229932; MAF= 0.02299%, 65/282692 alleles, homozygotes = 1) and has highest observed frequency in the Admixed American population (AF= 0.00177886; MAF= 0.17789%, 63/35416 alleles, homozygotes = 1); grpmax FAF= 0.00140947.
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.0055% · 89 / 1,613,924
1 hom · FAF 0.12%
Admixed American
86 / 59,994
0.14%
1 hom
Middle Eastern
1 / 6,084
0.016%
Remaining individuals
2 / 62,488
0.0032%
+ 7 not observed (European (Finnish), Amish, East Asian, South Asian, Ashkenazi Jewish, African/African American, European (non-Finnish))
gnomAD v2.1
0.023% · 65 / 282,692
1 hom · FAF 0.14%
Admixed American
63 / 35,416
0.18%
1 hom
Remaining individuals
2 / 7,218
0.028%
+ 6 not observed (African/African American, Ashkenazi Jewish, East Asian, European (Finnish), European (non-Finnish), South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Benign (4 clinical laboratories) and as Likely benign (4 clinical laboratories) and as Uncertain significance (1 clinical laboratory) and as Benign by Evidence-based Network for the Interpretation of Germline Mutant Alleles (ENIGMA) (expert panel). (ClinVarID = 52896)
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.01).
Functional / OncoKB screenshot
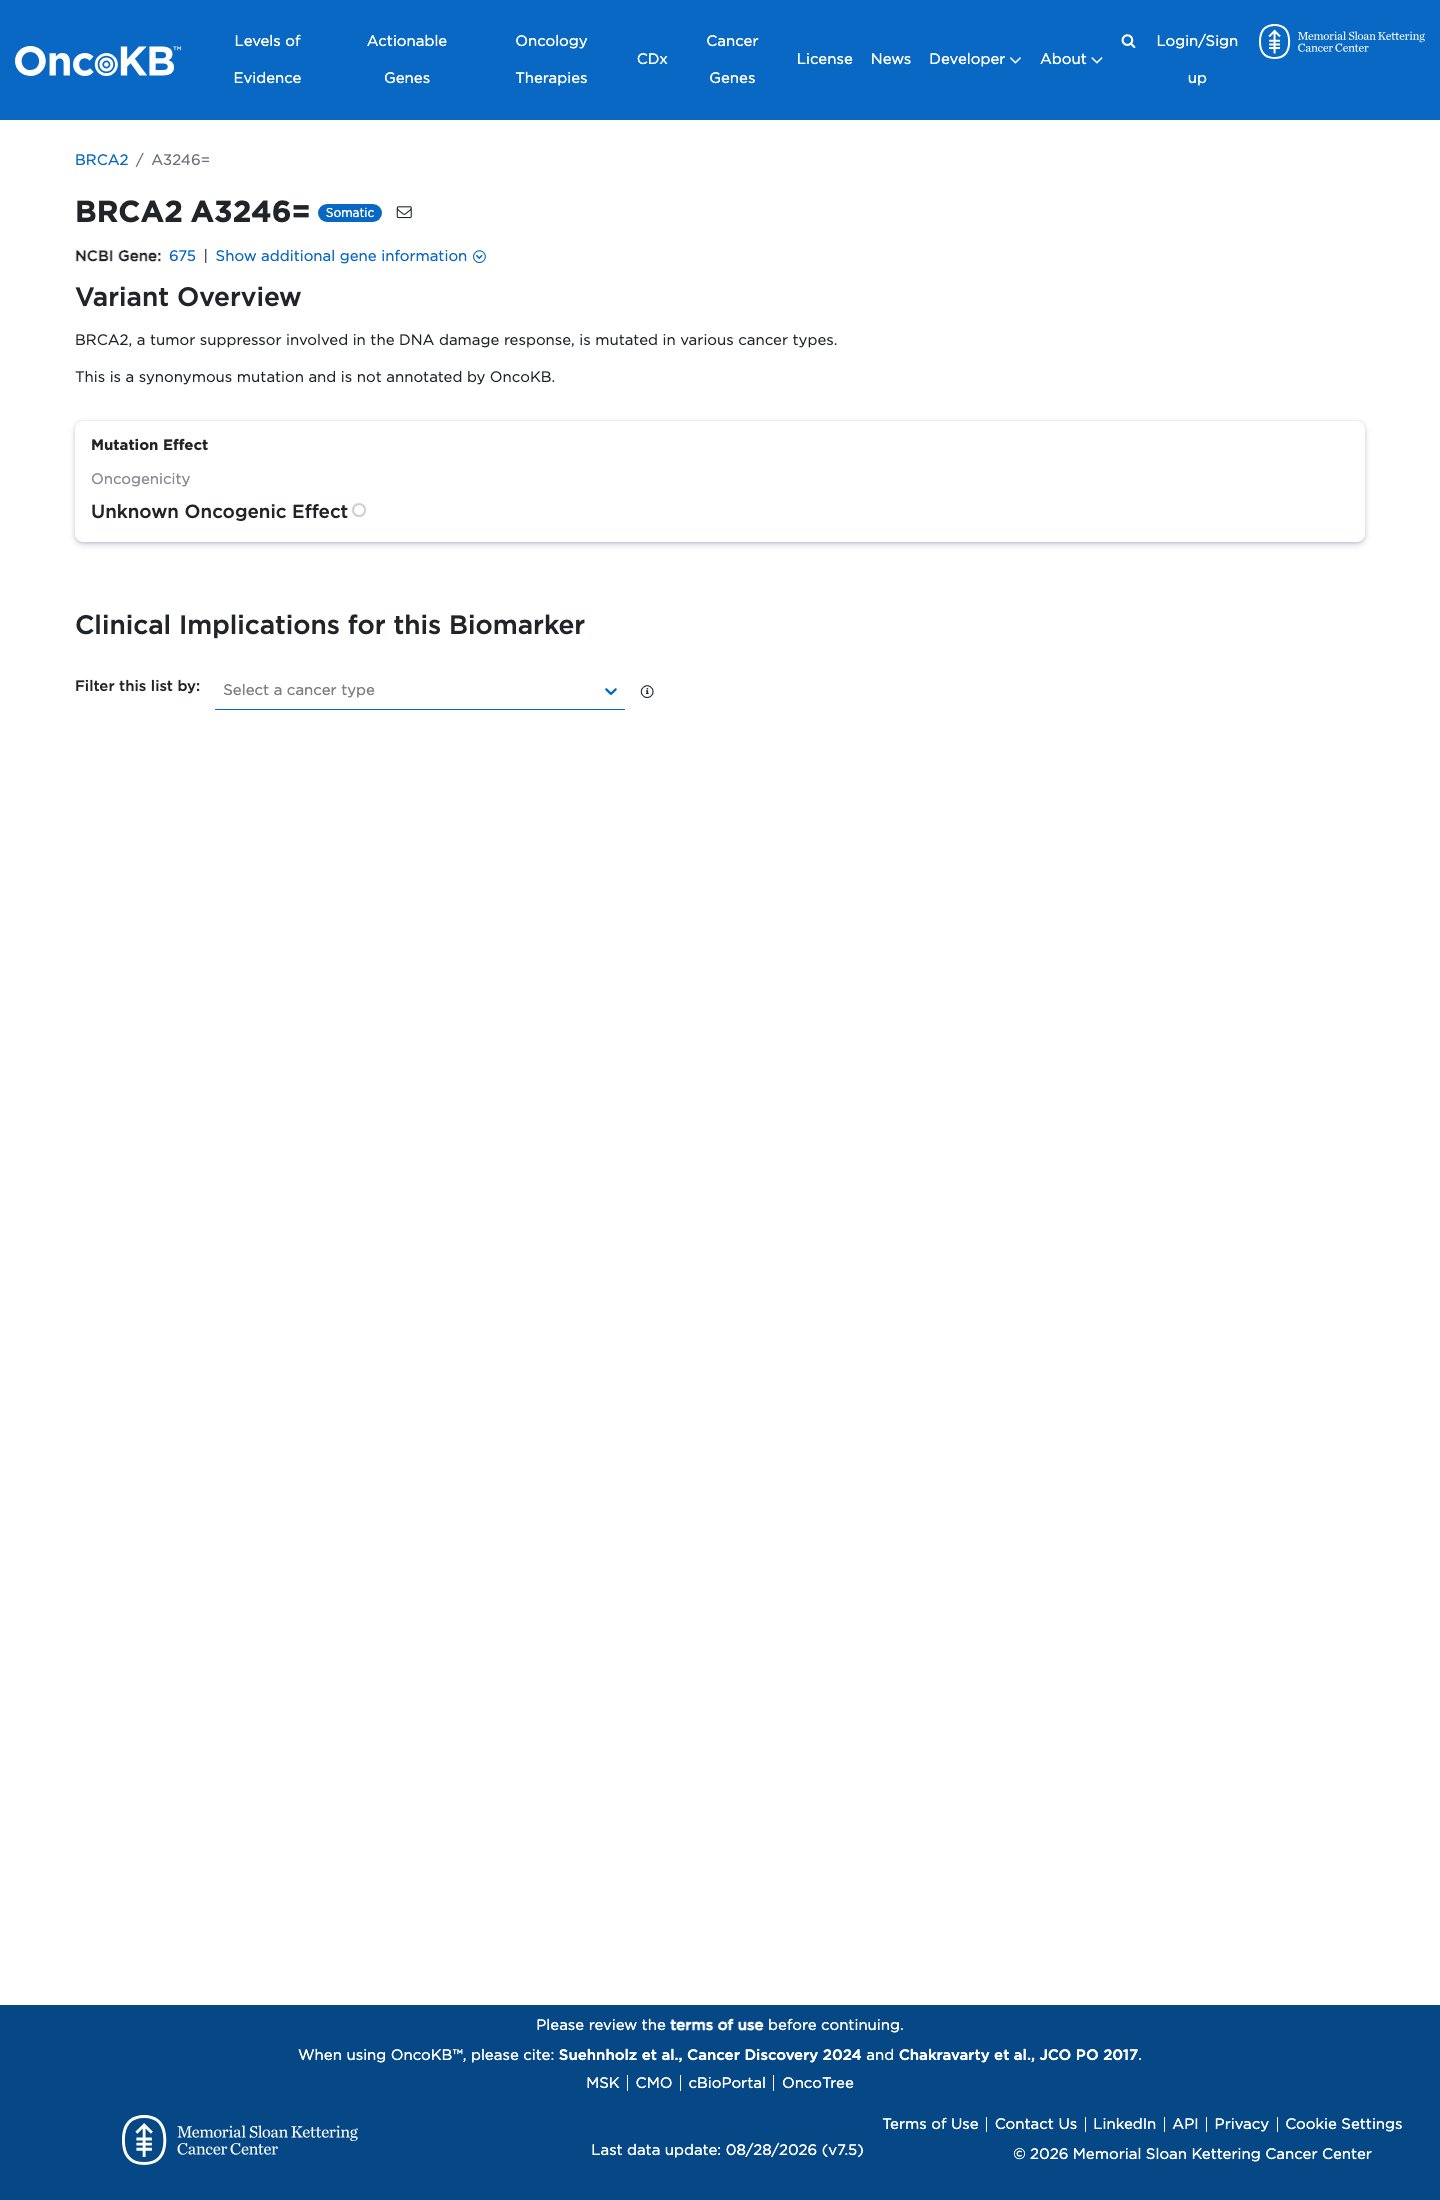
Functional Unknown Oncogenic Effect
OncoKB identified curated literature and non-variant-specific oncogenicity context for review; listed oncogenicity label: Unknown Oncogenic Effect.
OncoKB ↗
COSMIC screenshot
COSMIC
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
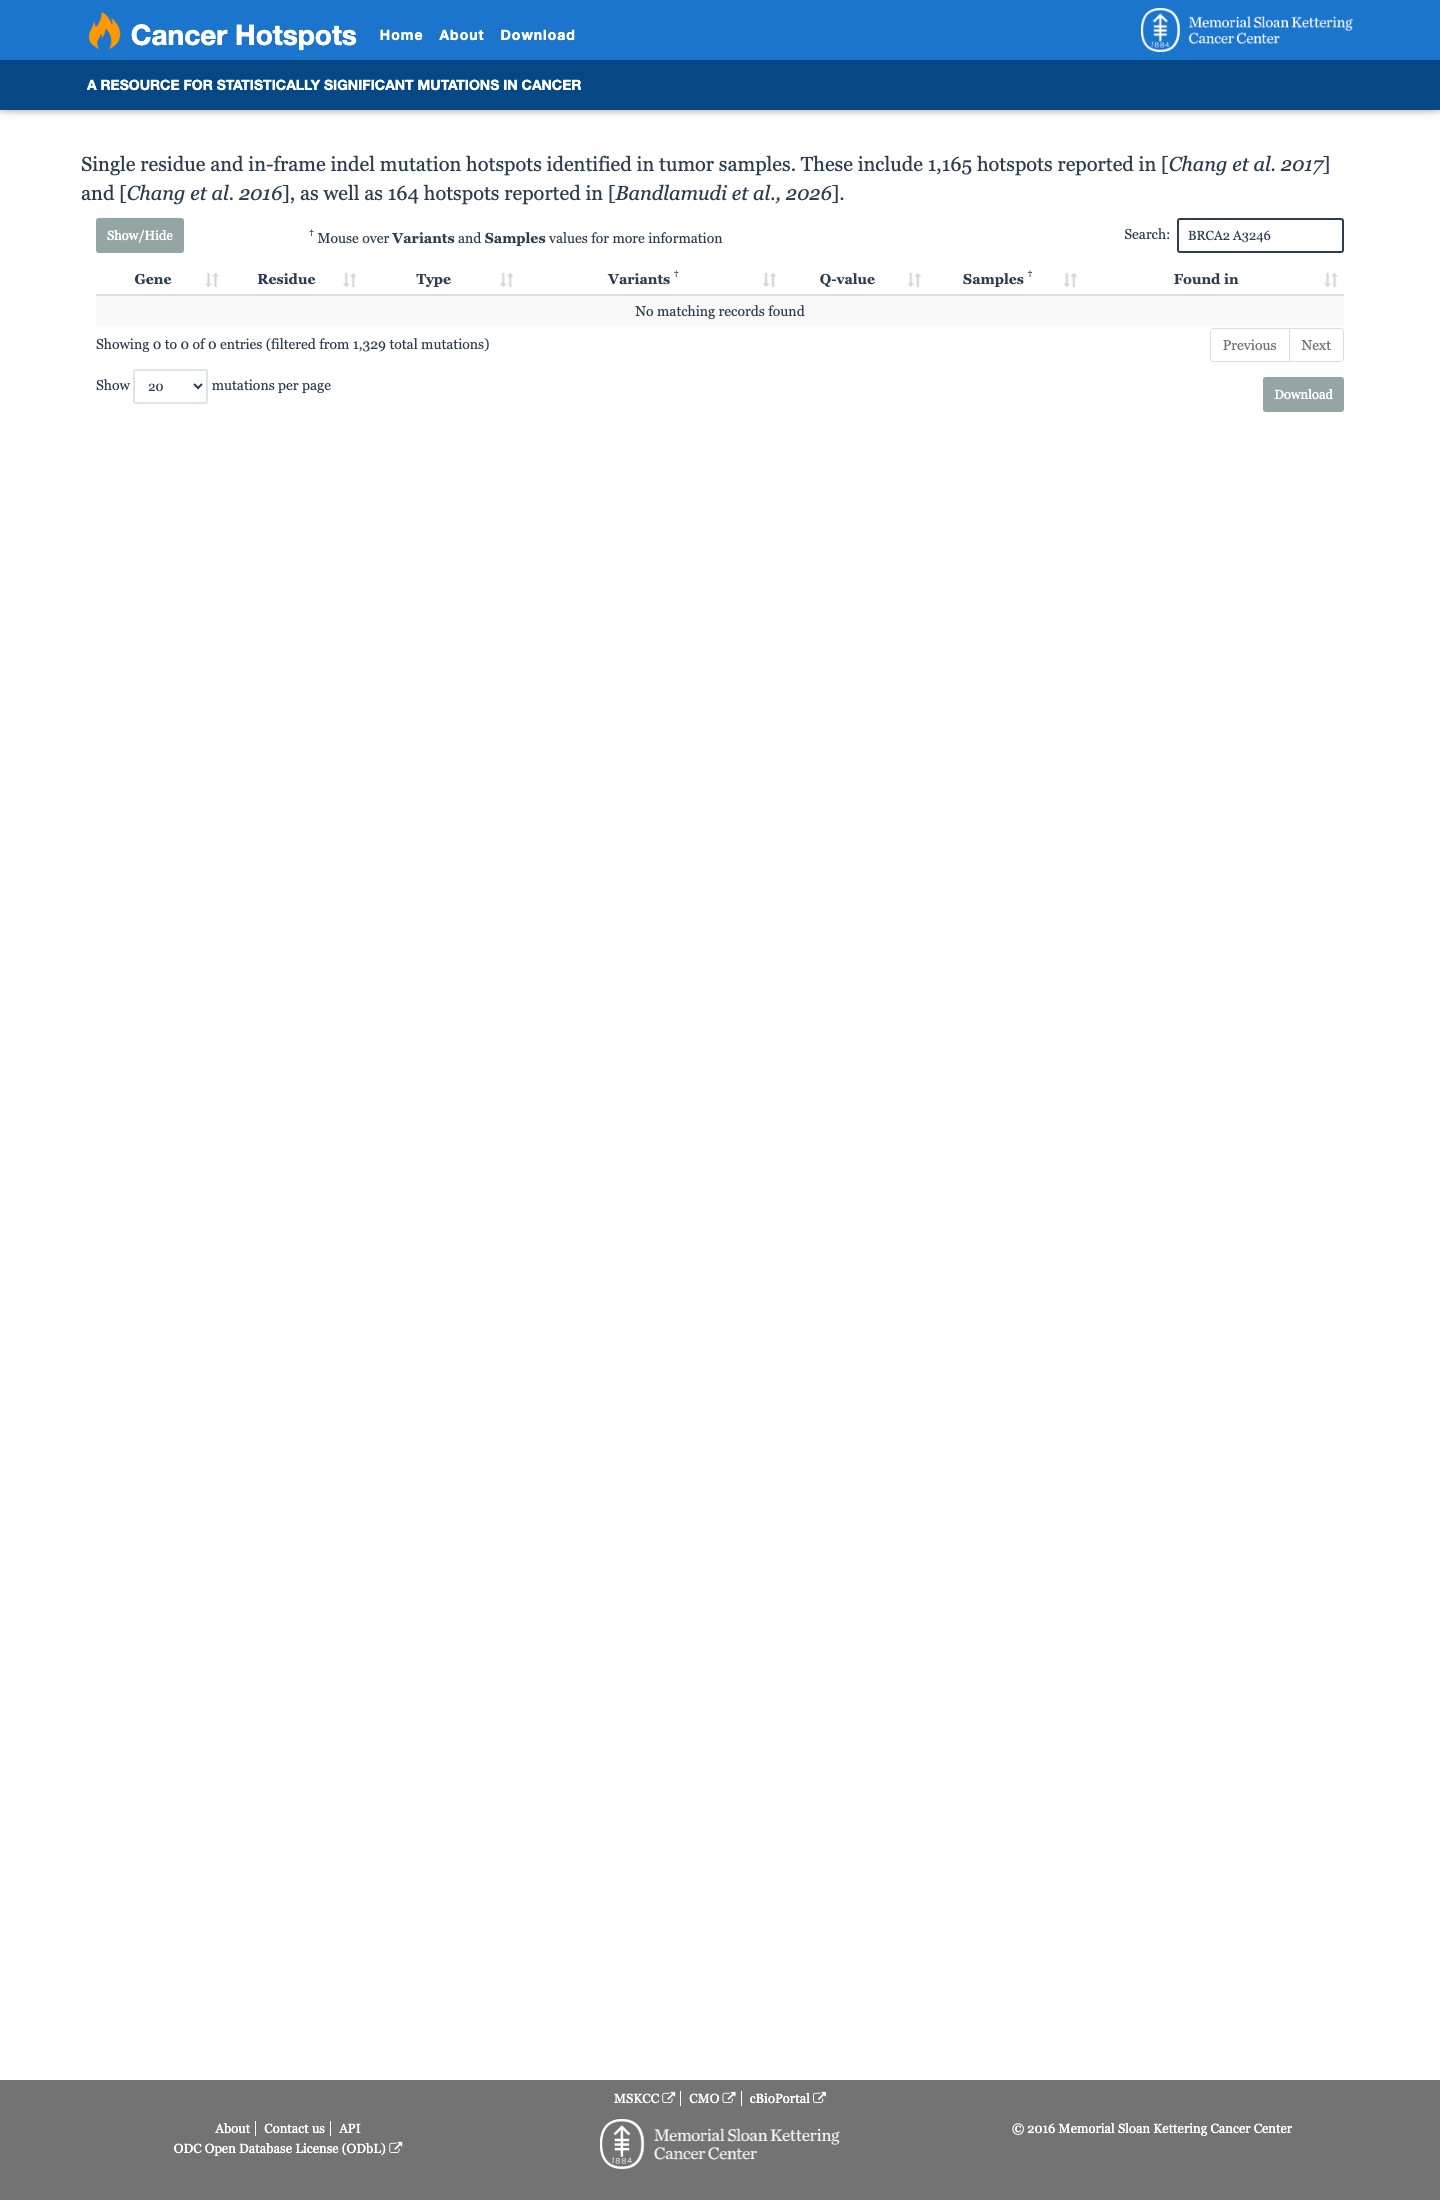
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 9 PMIDs not cited in assessment
23918944 ↗ Tamoxifen and risk of contralateral breast cancer for BRCA1 and BRCA2 mutation carriers. CLINVAR
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology. CLINVAR
34242744 ↗ Customizing local and systemic therapies for women with early breast cancer: the St. Gallen International Consensus Guidelines for treatment of early breast cancer 2021. CLINVAR
12442275 ↗ BRCA1 and BRCA2 mutation analysis of early-onset and familial breast cancer cases in Mexico. CLINVAR
15889636 ↗ Clinical follow up of mexican women with early onset of breast cancer and mutations in the BRCA1 and BRCA2 genes. CLINVAR
25356965 ↗ ACMG policy statement: updated recommendations regarding analysis and reporting of secondary findings in clinical genome-scale sequencing. CLINVAR
26389258 ↗ Cancer Genetics Risk Assessment and Counseling (PDQ®): Health Professional Versi CLINVAR
34012068 ↗ ACMG SF v3.0 list for reporting of secondary findings in clinical exome and genome sequencing: a policy statement of the American College of Medical Genetics and Genomics (ACMG). CLINVAR
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification criteria. CLINVAR