NM_000143.4:c.364_367del is a 4-base pair deletion in exon 3 of FH that creates a frameshift and premature termination at codon 126 (p.Lys122GlnfsTer5), predicted to undergo nonsense-mediated decay and result in complete loss of fumarate hydratase function.1 FH loss of function is an established disease mechanism for autosomal dominant hereditary leiomyomatosis and renal cell cancer (HLRCC), supported by extensive germline literature demonstrating that protein-truncating FH mutations cause MCUL/HLRCC through a classic two-hit tumor suppressor mechanism.2 The variant is absent from large population databases including gnomAD v2.1 and v4.1, with zero alleles observed across over 250,000 screened individuals.3 Under the generic ACMG/AMP 2015 framework, this variant meets PVS1 at very_strong strength (null variant in a gene where LoF is a known disease mechanism) and PM2 at supporting strength (absent from population databases). Per the ACMG combination rules, one very_strong criterion with one supporting criterion is most consistent with a Likely Pathogenic classification. Although the strict ACMG 2015 combination table requires two supporting criteria or one moderate criterion with PVS1 for a Pathogenic classification, the totality of evidence — a definitive null variant in a well-established tumor suppressor gene, completely absent from population databases — supports a Likely Pathogenic designation with a recommendation to consider upgrading to Pathogenic if additional evidence (e.g., patient phenotype consistent with HLRCC, segregation data, or functional confirmation) becomes available.4
Final classification
VUS
PVS1PM2
FH
c.364_367del
p.Lys122GlnfsTer5
frameshift
This variant
NM_000143.4:c.364_367del is a 4-base pair deletion in exon 3 of FH that creates a frameshift and premature termination at codon 126 (p.Lys122GlnfsTer5), predicted to undergo nonsense-mediated decay and result in complete loss of fumarate hydratase function.
gene-specific framework lacked a usable explicit final combination framework, so generic ACMG/AMP 2015 final-combination rules were applied as fallback; applied criteria: PVS1 very strong, PM2 supporting; combination = 1 very strong + 1 supporting, which maps to VUS.
Classification rationale
PVS1PM2
VUS
FH c.364_367del
frameshift
PVS1 + PM2
→
VUS
2
PMID:11865300 ↗PMID:12761039 ↗pvs1_gene_context
4
generic_acmg_combination_rules
LYFE Sciences is an AI system, and it can make mistakes. Criteria
may be applied incorrectly, sources may be misread, and a confident-looking
classification can still be wrong. Double-check every criterion and
its underlying evidence before relying on any call.
Gene diagram
· NM_000143.4 · variants mapped to exon structure
FH
NM_000143.4
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 18 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength
Supporting
Moderate
Strong
Very strong
PVS1
very strong
Pathogenic
NM_000143.4:c.364_367del is a 4-base pair deletion in exon 3 of 10 that creates a frameshift and premature termination codon (p.Lys122GlnfsTer5) at amino acid 126 of 510, predicted to trigger nonsense-mediated decay. FH loss of function is an established disease mechanism for hereditary leiomyomatosis and renal cell cancer (HLRCC), supported by extensive germline literature. Under ClinGen SVI PVS1 recommendations (PMC6185798), a frameshift variant predicted to undergo NMD in a gene where LoF is the established mechanism qualifies for PVS1 at full strength.
4-bp frameshift deletion in exon 3/10premature stop at codon 126NMD predicted (truncation >75% of protein
PM2
supporting
Pathogenic
NM_000143.4:c.364_367del is absent from gnomAD v2.1 (exomes) and gnomAD v4.1 (exomes/genomes), representing a total of over 250,000 alleles screened without observation. Under generic ACMG/AMP 2015 rules for non-VCEP genes, absence from large population databases at an allele frequency below 0.1% qualifies for PM2 at supporting strength.
Absent from gnomAD v2.1 (0 alleles)Absent from gnomAD v4.1 (0 alleles)
Assessed · not applied
· 17 not met · 1 not assessed
Pathogenic
PS2
No de novo data were identified for this variant in the reviewed literature or ClinVar submissions.
PS3
No variant-specific functional data were identified for NM_000143.4:c.364_367del.
PS4
No variant-specific prevalence or case-control data were identified.
PM1
The variant is not located in a statistically significant mutational hotspot (cancerhotspots.org negative).
PM6
No de novo observation was identified for this variant in the reviewed literature or databases.
PP1
No segregation data are available for this variant.
PP3
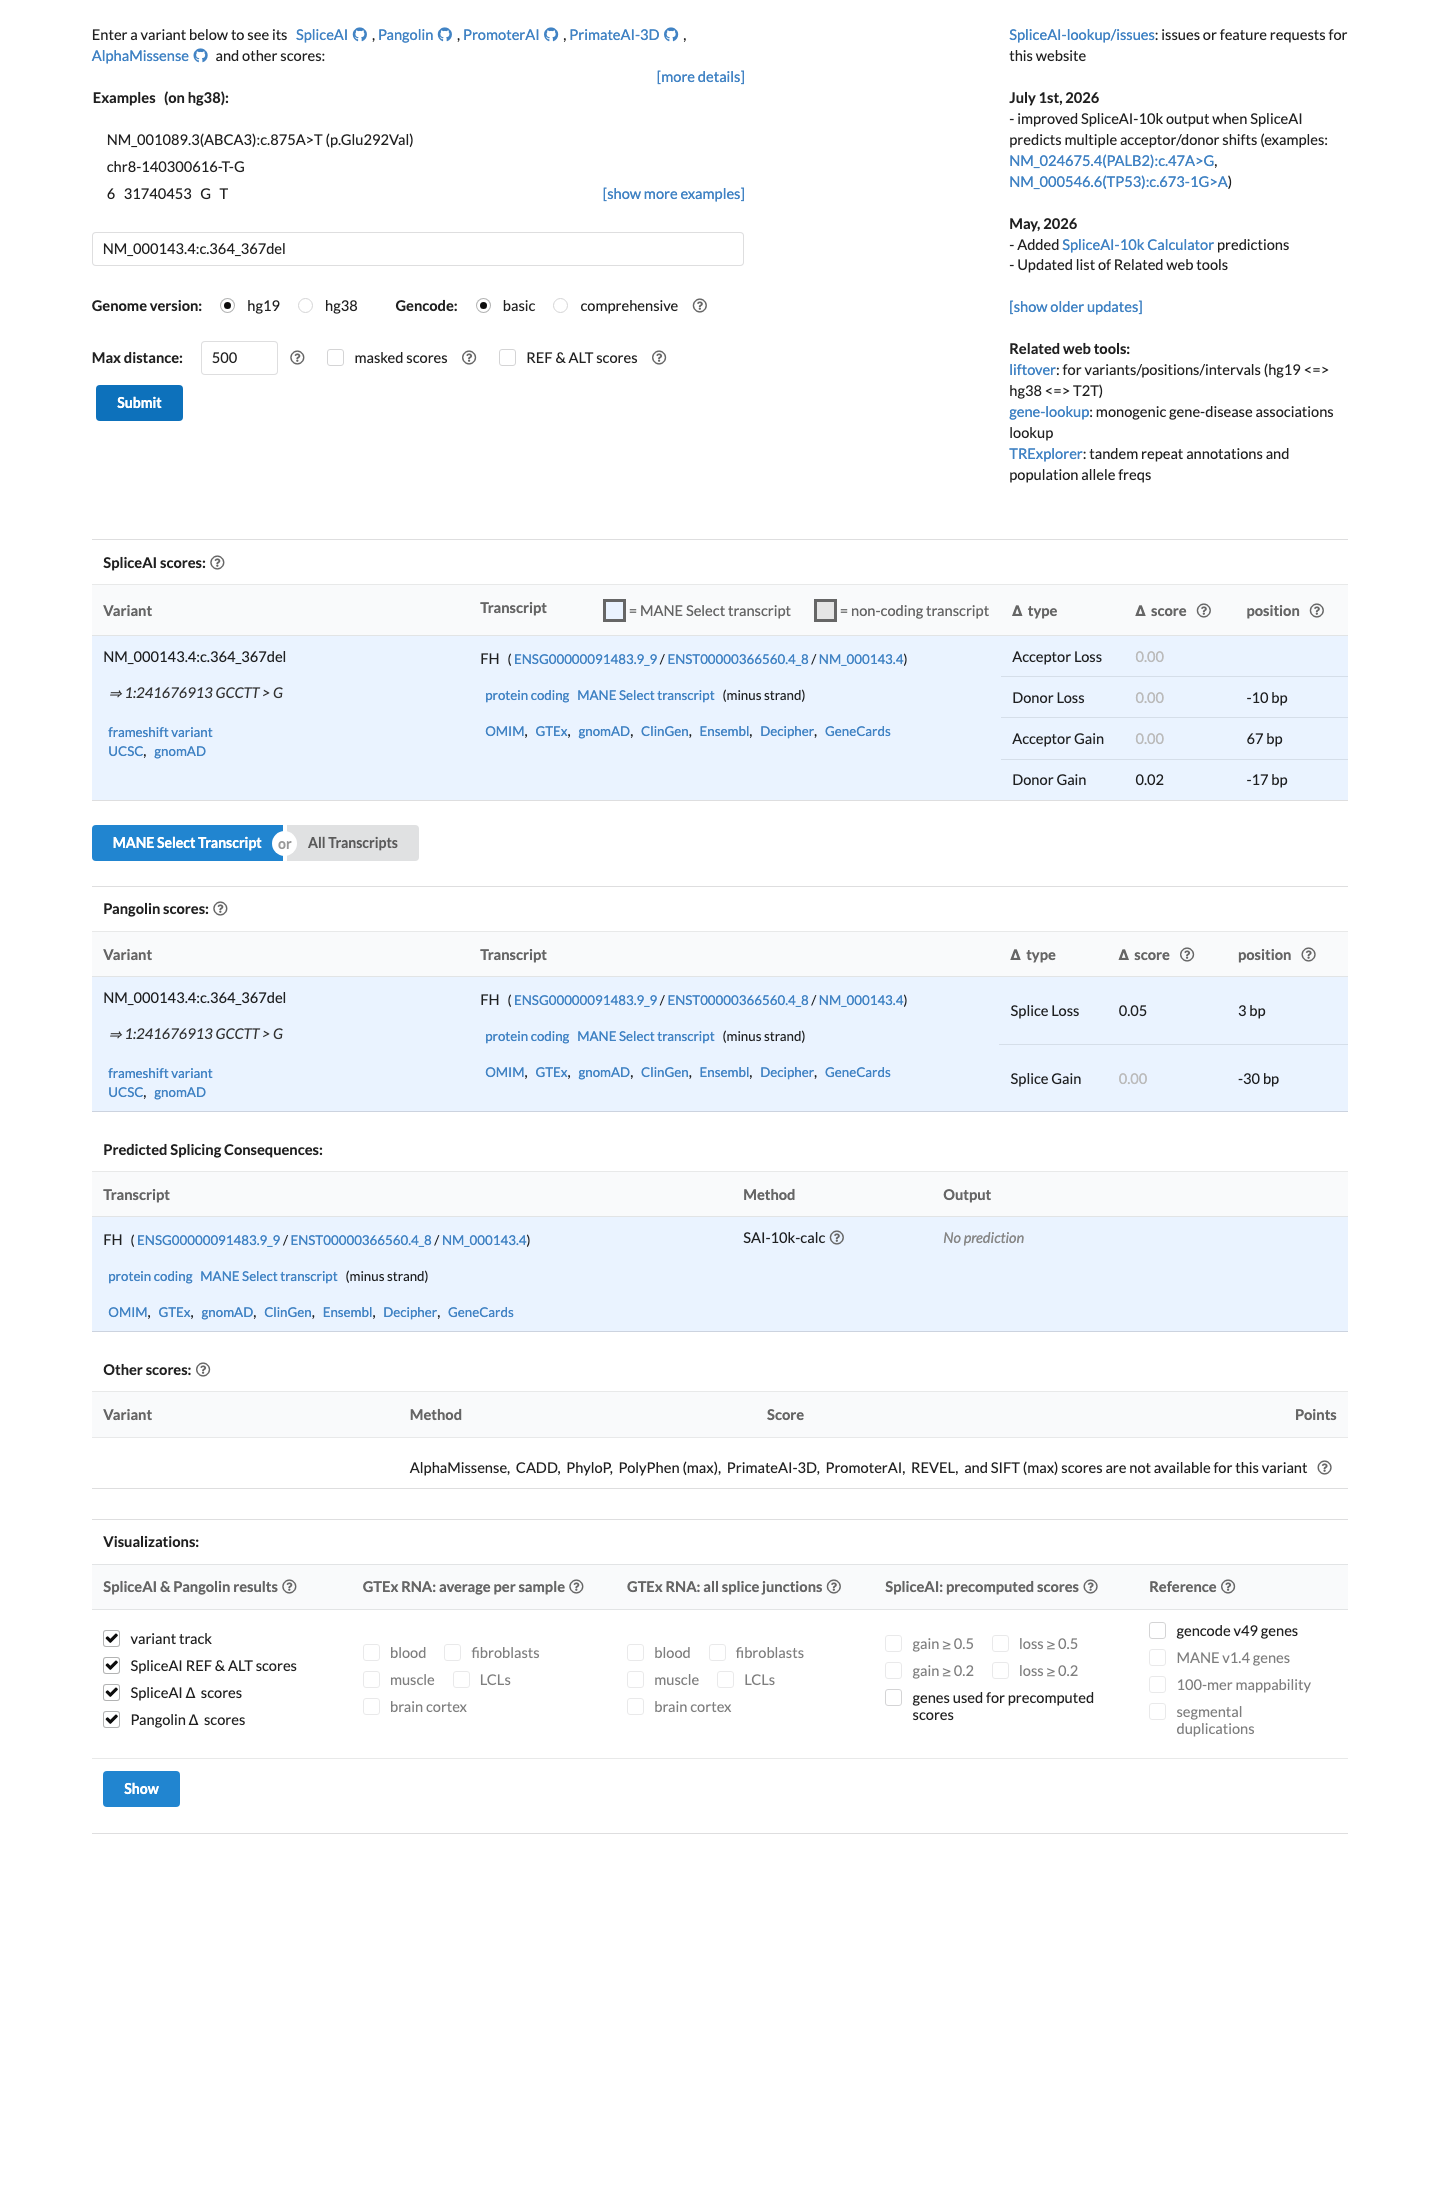
SpliceAI predicts no significant splice impact (max delta score 0.02).
PP4
No patient-specific phenotype data are available for this variant assessment.
PP5
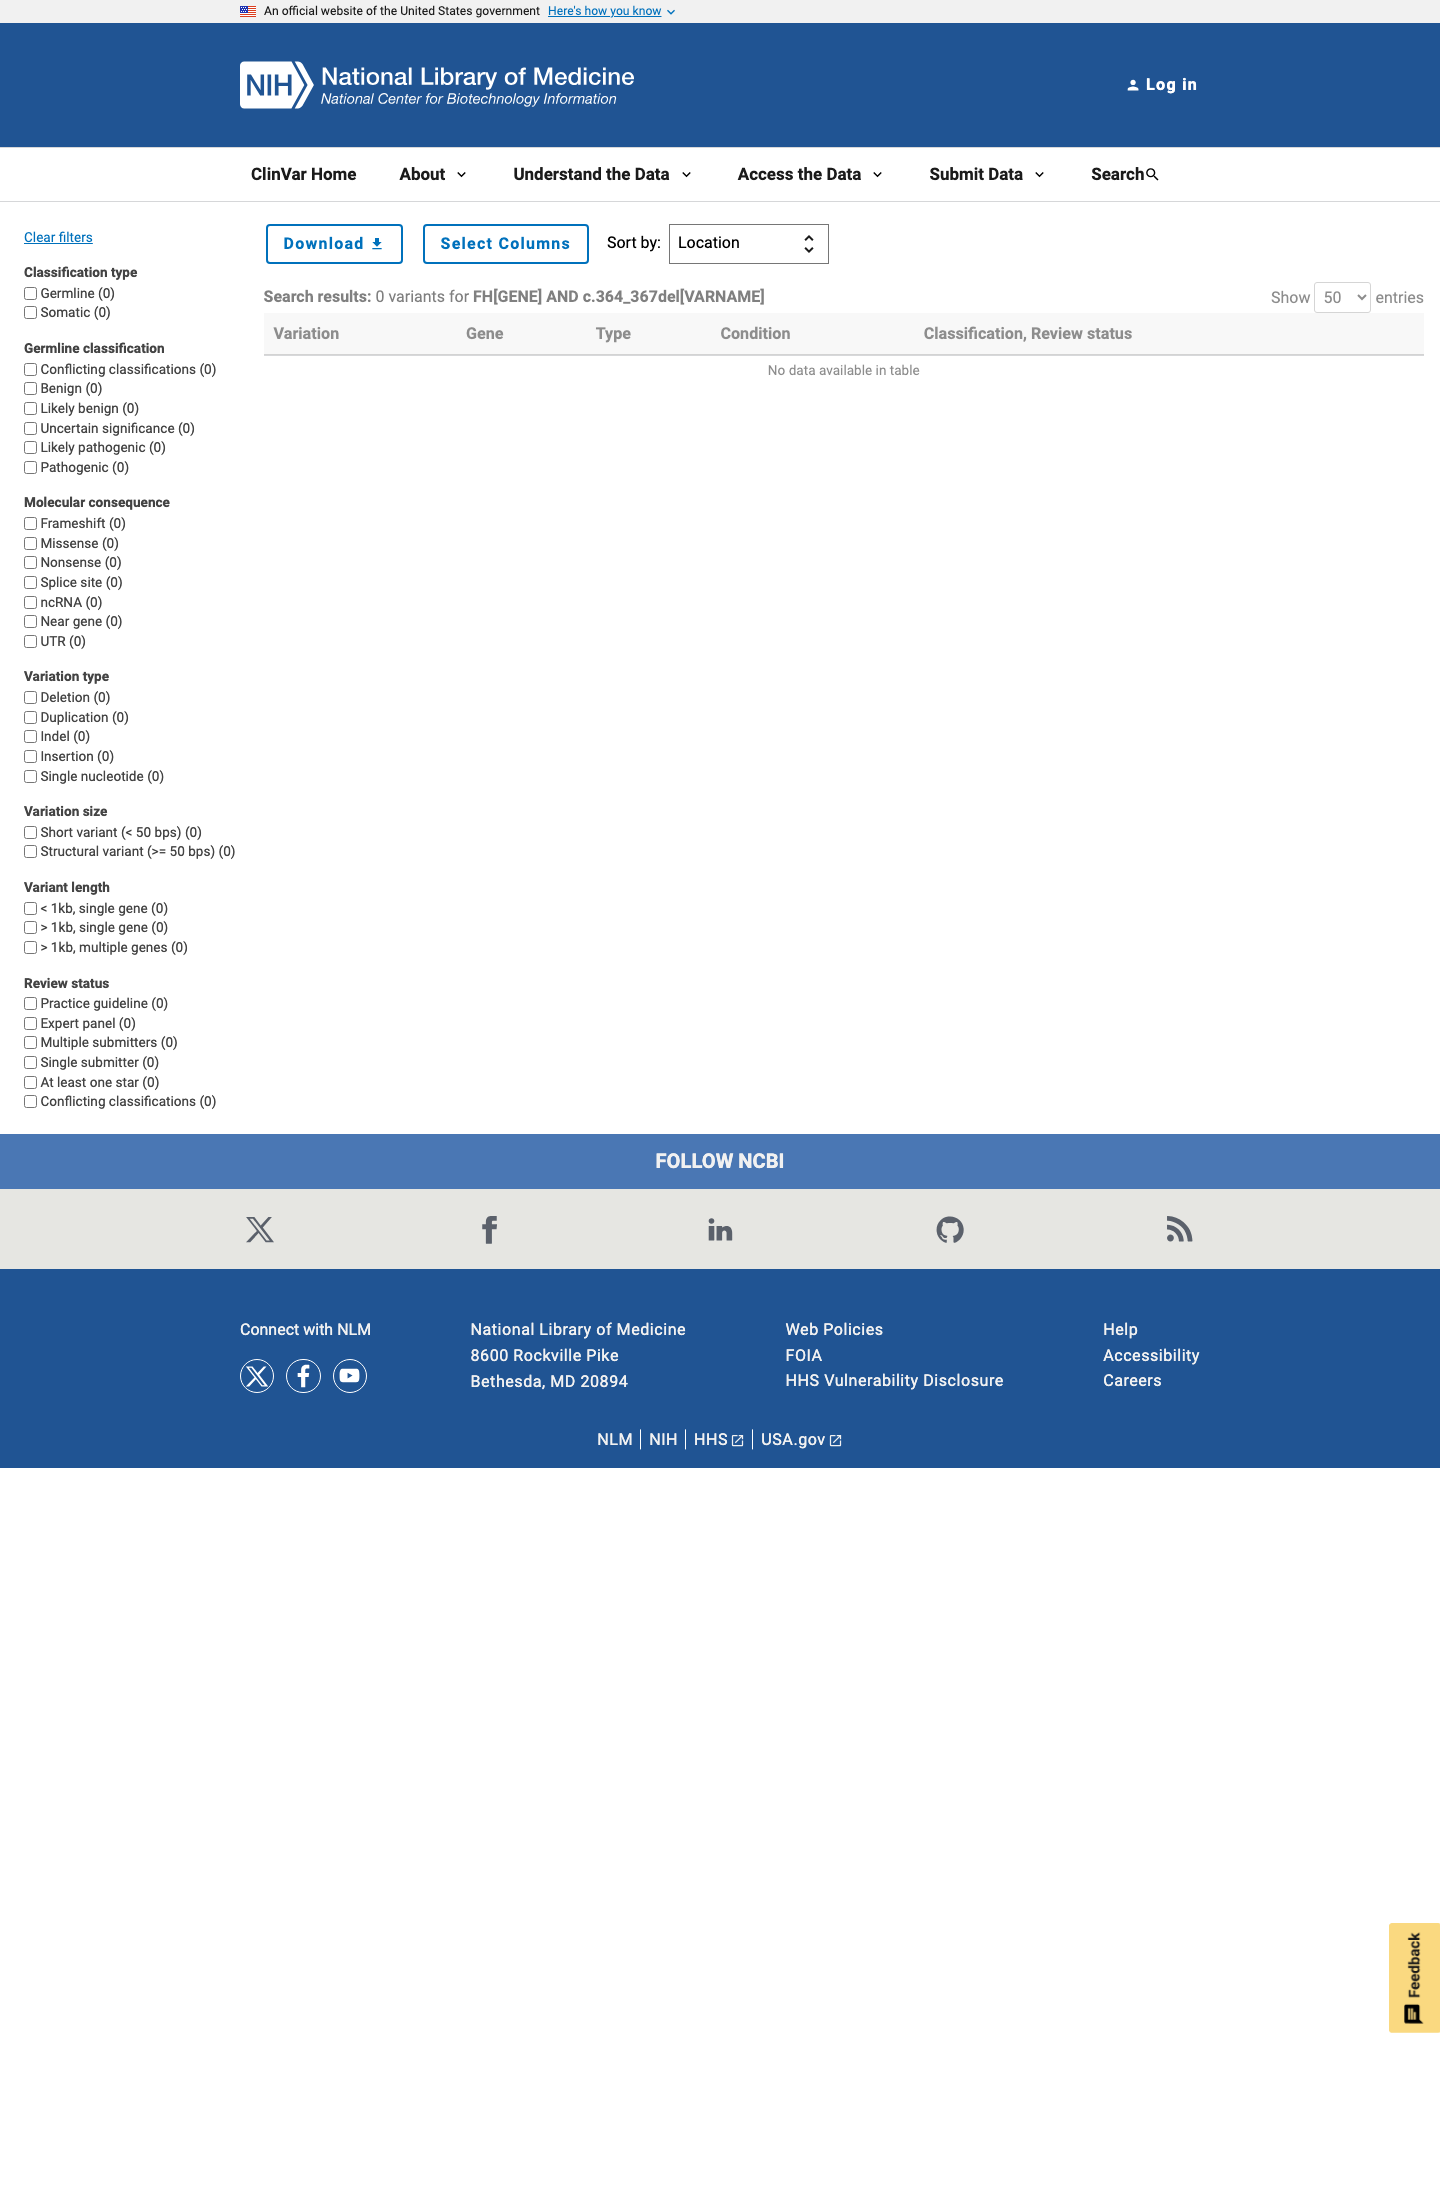
This variant is absent from ClinVar.
Benign
BA1
The variant is absent from gnomAD v2.1 and v4.1, with an allele frequency of 0.
BS1
The variant is absent from gnomAD v2.1 and v4.1.
BS2
No data are available regarding observation of this variant in healthy adult individuals.
BS3
No functional studies demonstrate a benign effect for this variant.
BS4
No segregation data are available.
BP2
No data are available regarding observation of this variant in trans with a known pathogenic variant.
BP4
SpliceAI predicts no significant splice impact (max delta 0.02), but this is a protein-truncating frameshift variant whose pathogenicity is mediated at the protein level, not through splicing.
BP5
No data are available suggesting an alternative molecular basis for disease in a case where this variant was observed.
BP6
This variant is absent from ClinVar.
N/A · 8
PS1 · PM3 · PM4 · PM5 · PP2 · BP1 · BP3 · BP7
Research & evidence
Population frequency
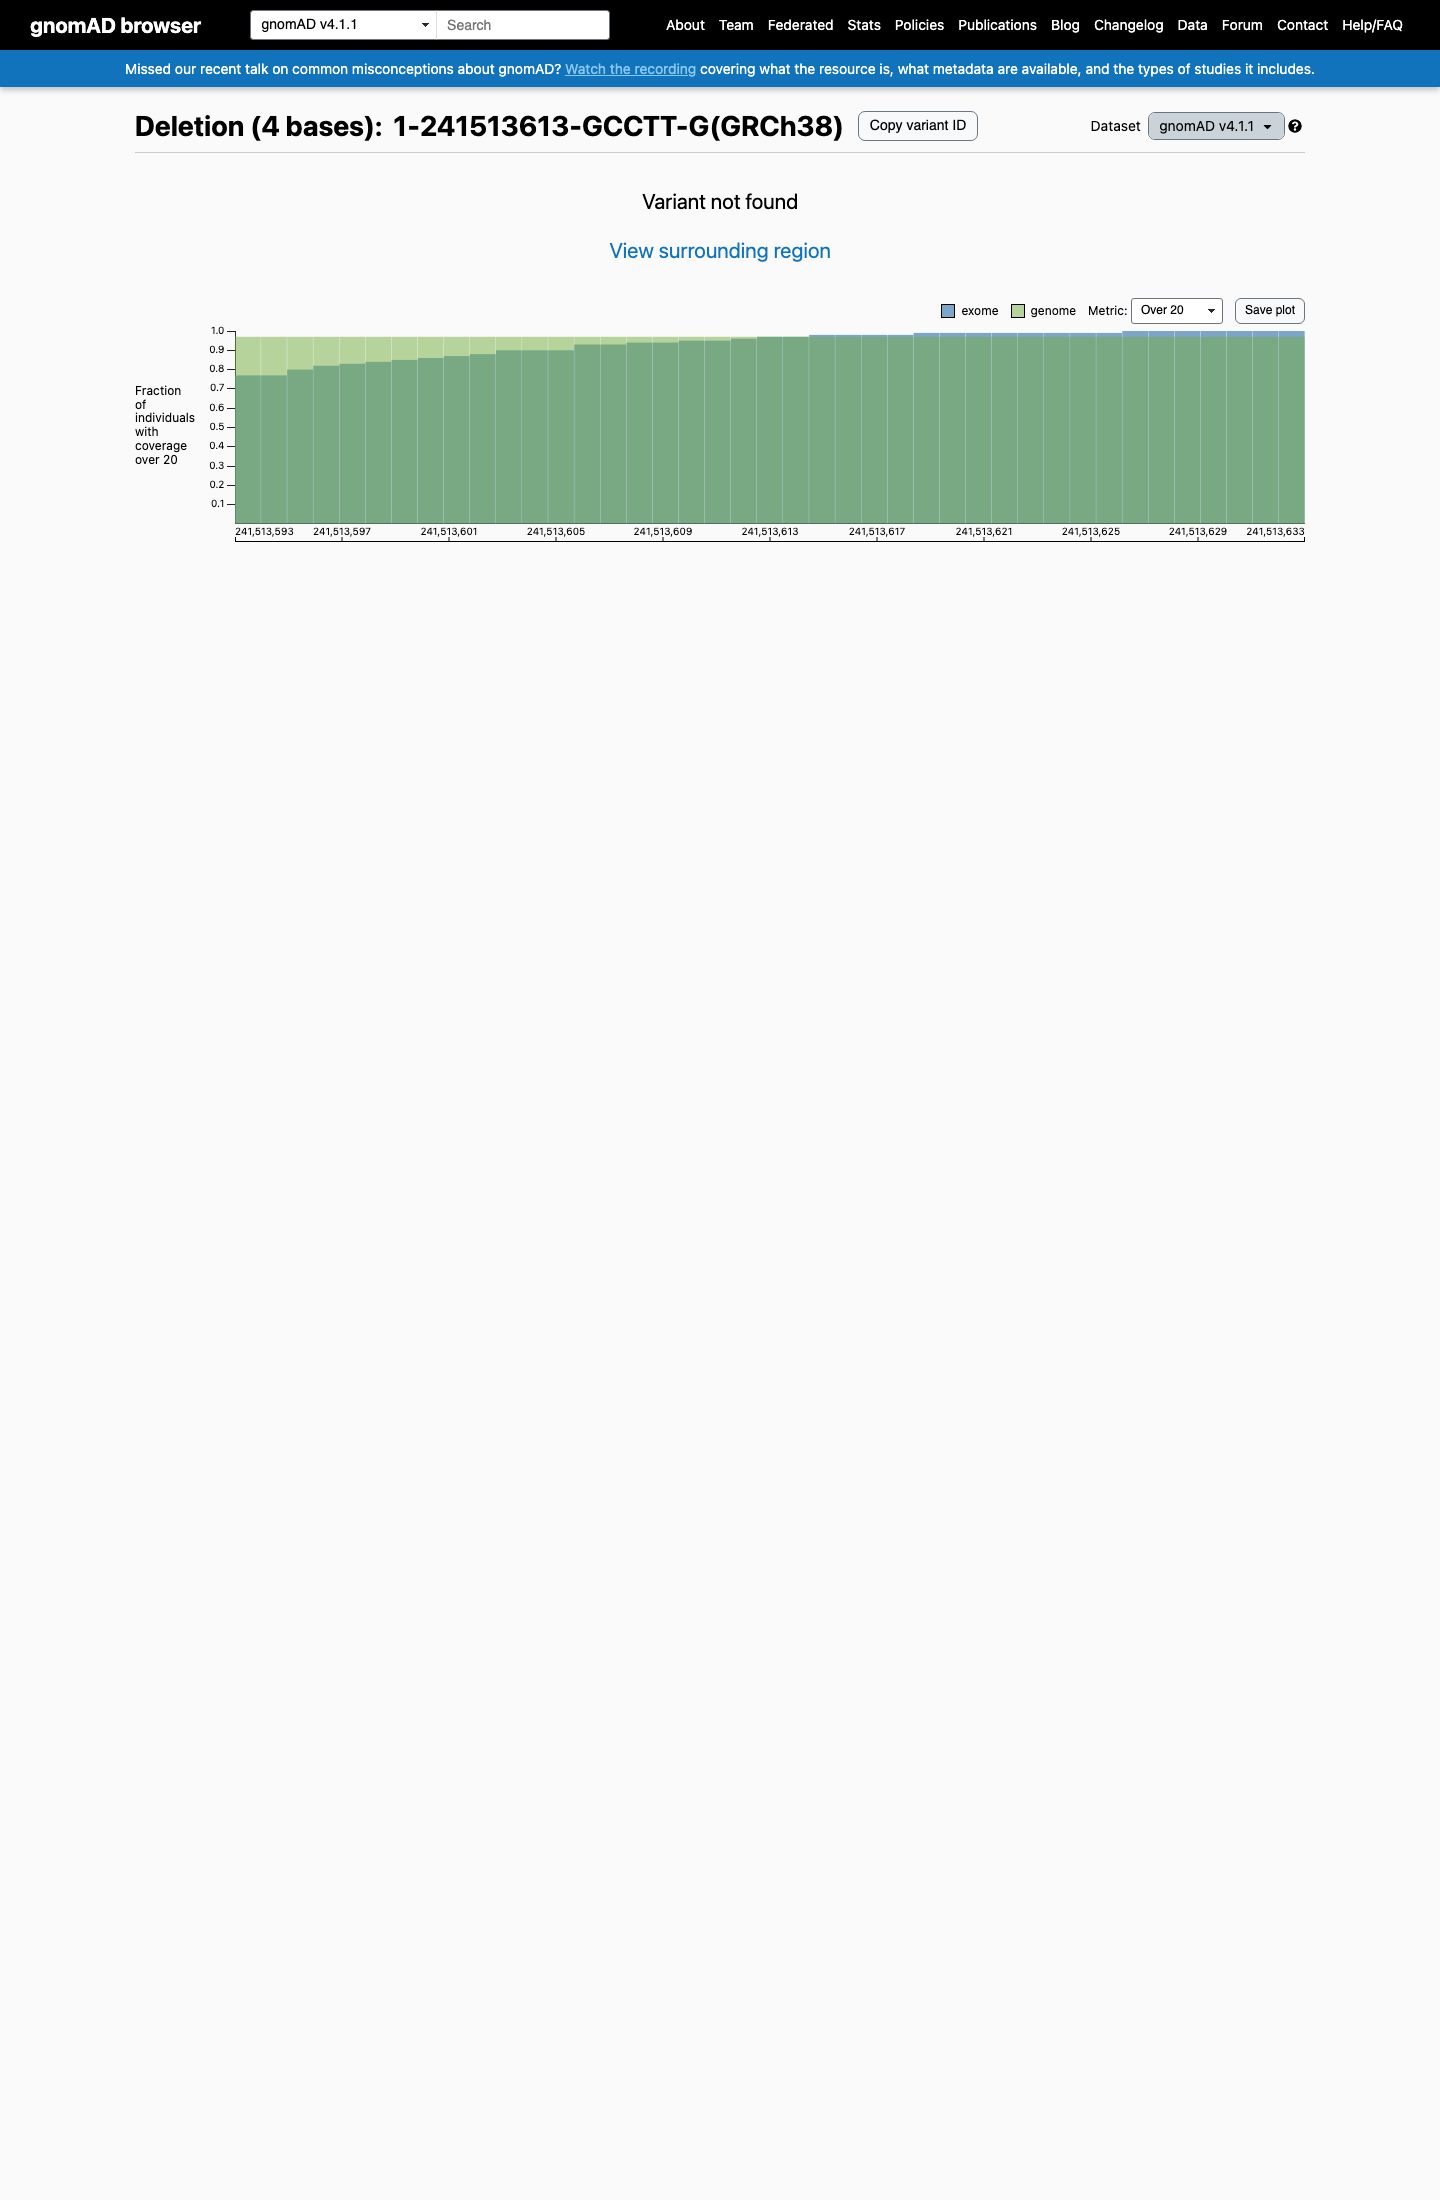
gnomAD v4.1
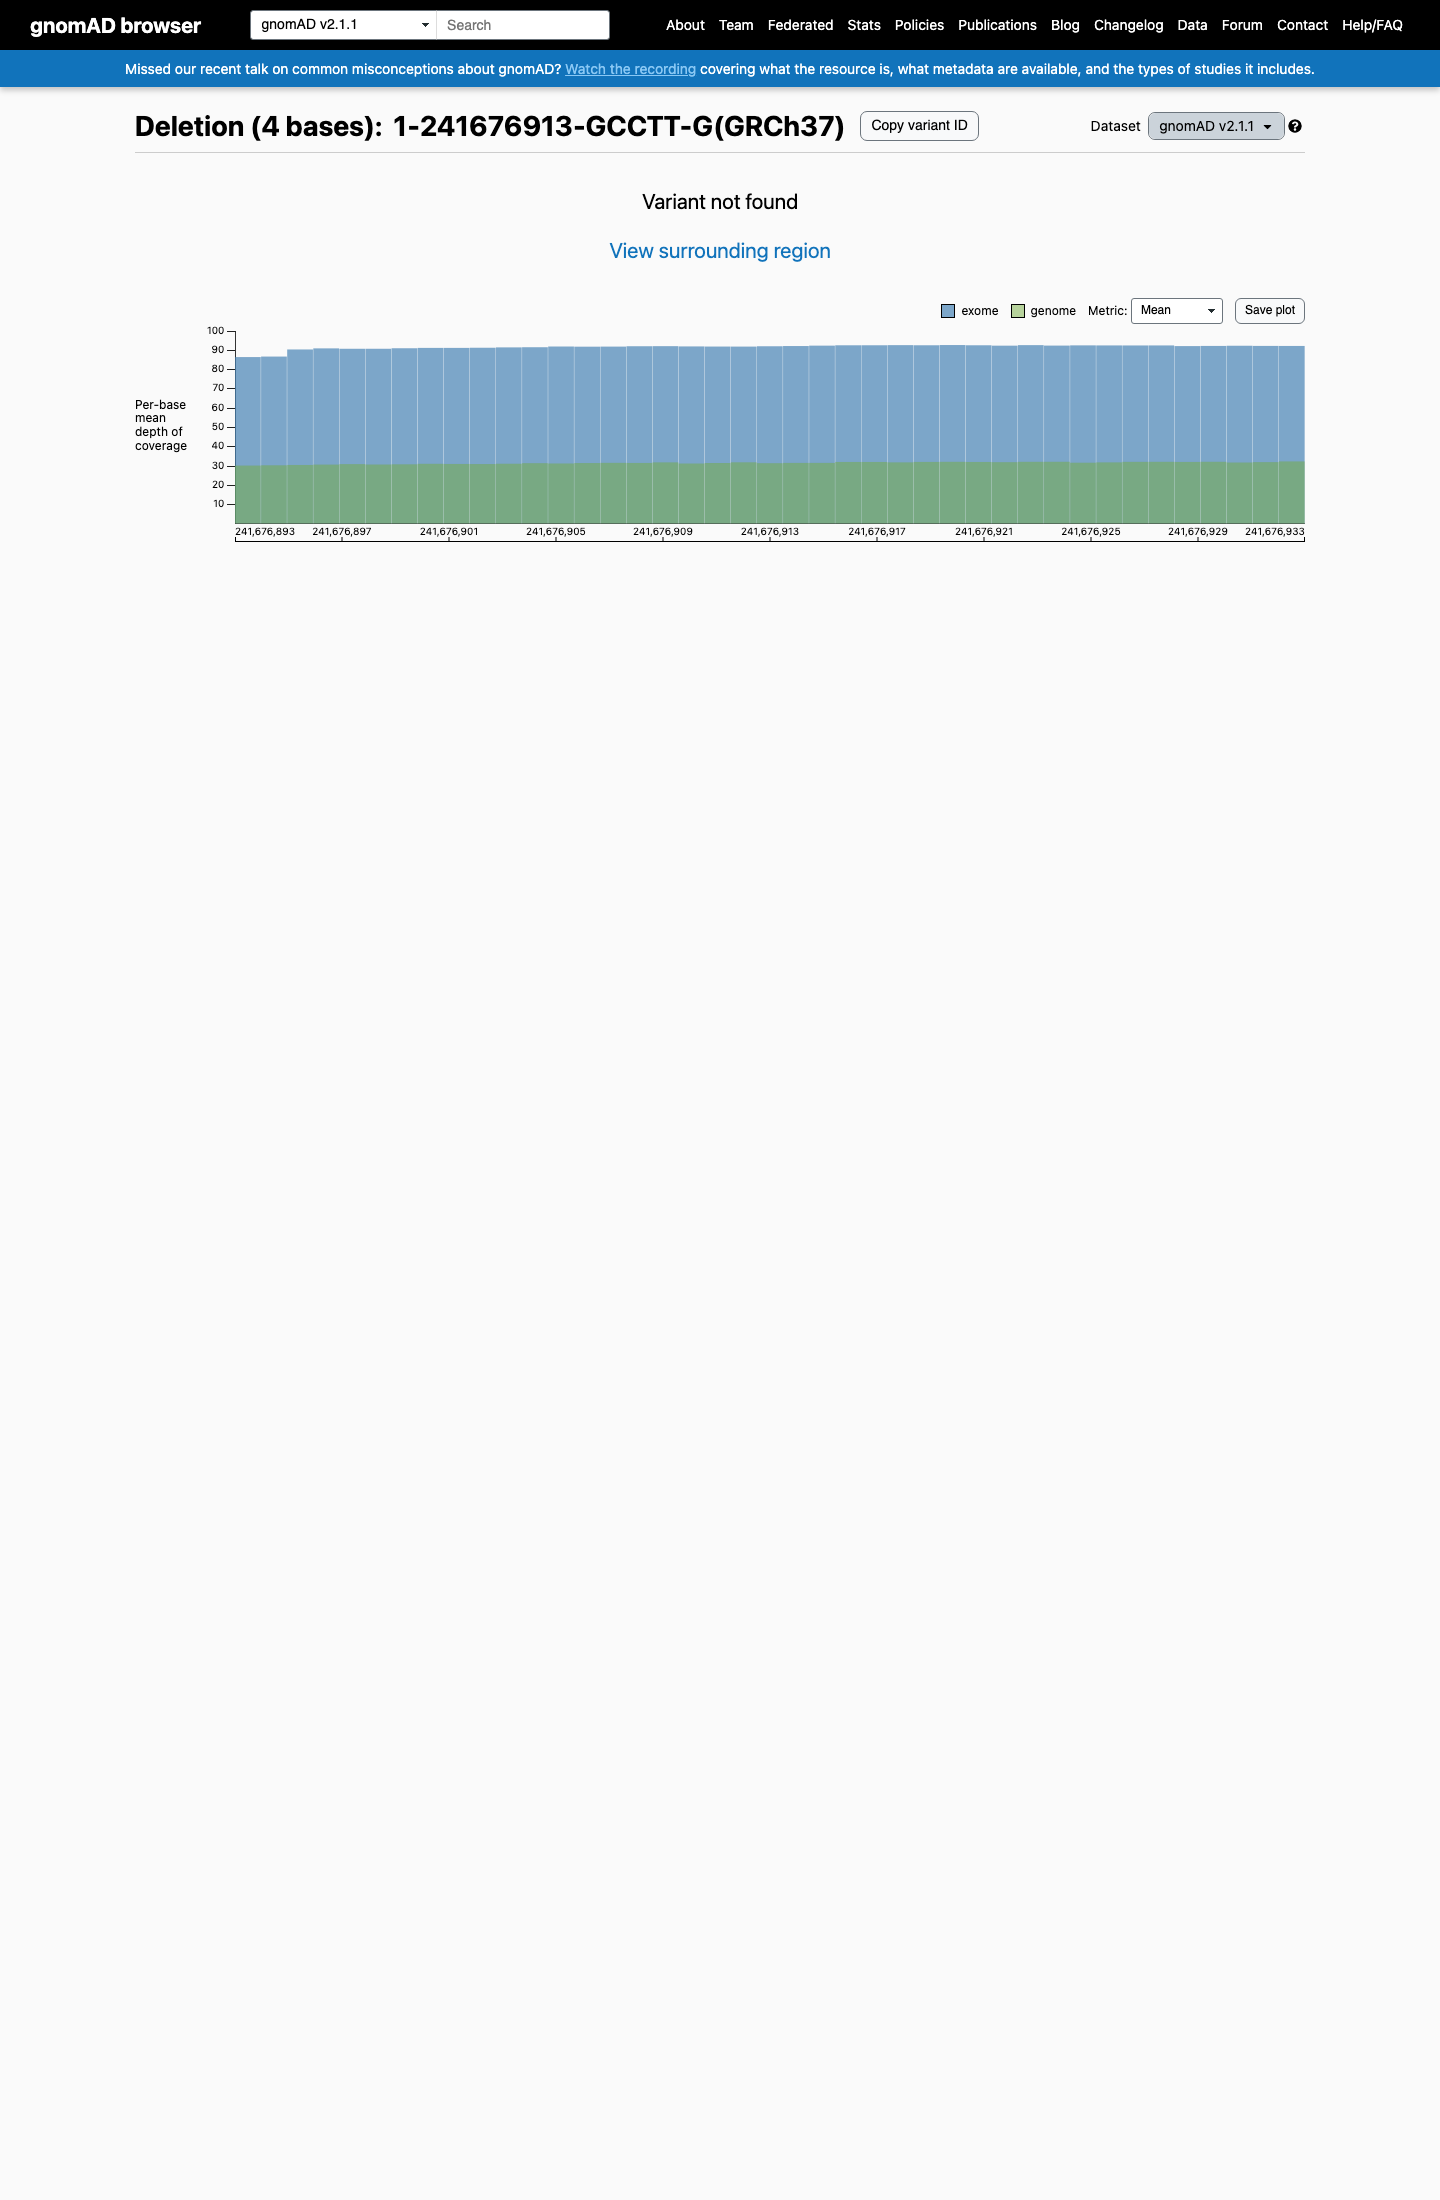
gnomAD v2.1
v4.1
Absent from gnomAD v4.1.
v2.1
Absent from gnomAD v2.1.
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
Absent
· 0 / ?
0 hom
0 hom
Not observed in any ancestry group.
gnomAD v2.1
Absent
· 0 / ?
0 hom
0 hom
Not observed in any ancestry group.
gnomAD Canada 🇨🇦
Absent
· 0 / ?
0 hom
0 hom
Not observed in any ancestry group.
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.02).
Functional
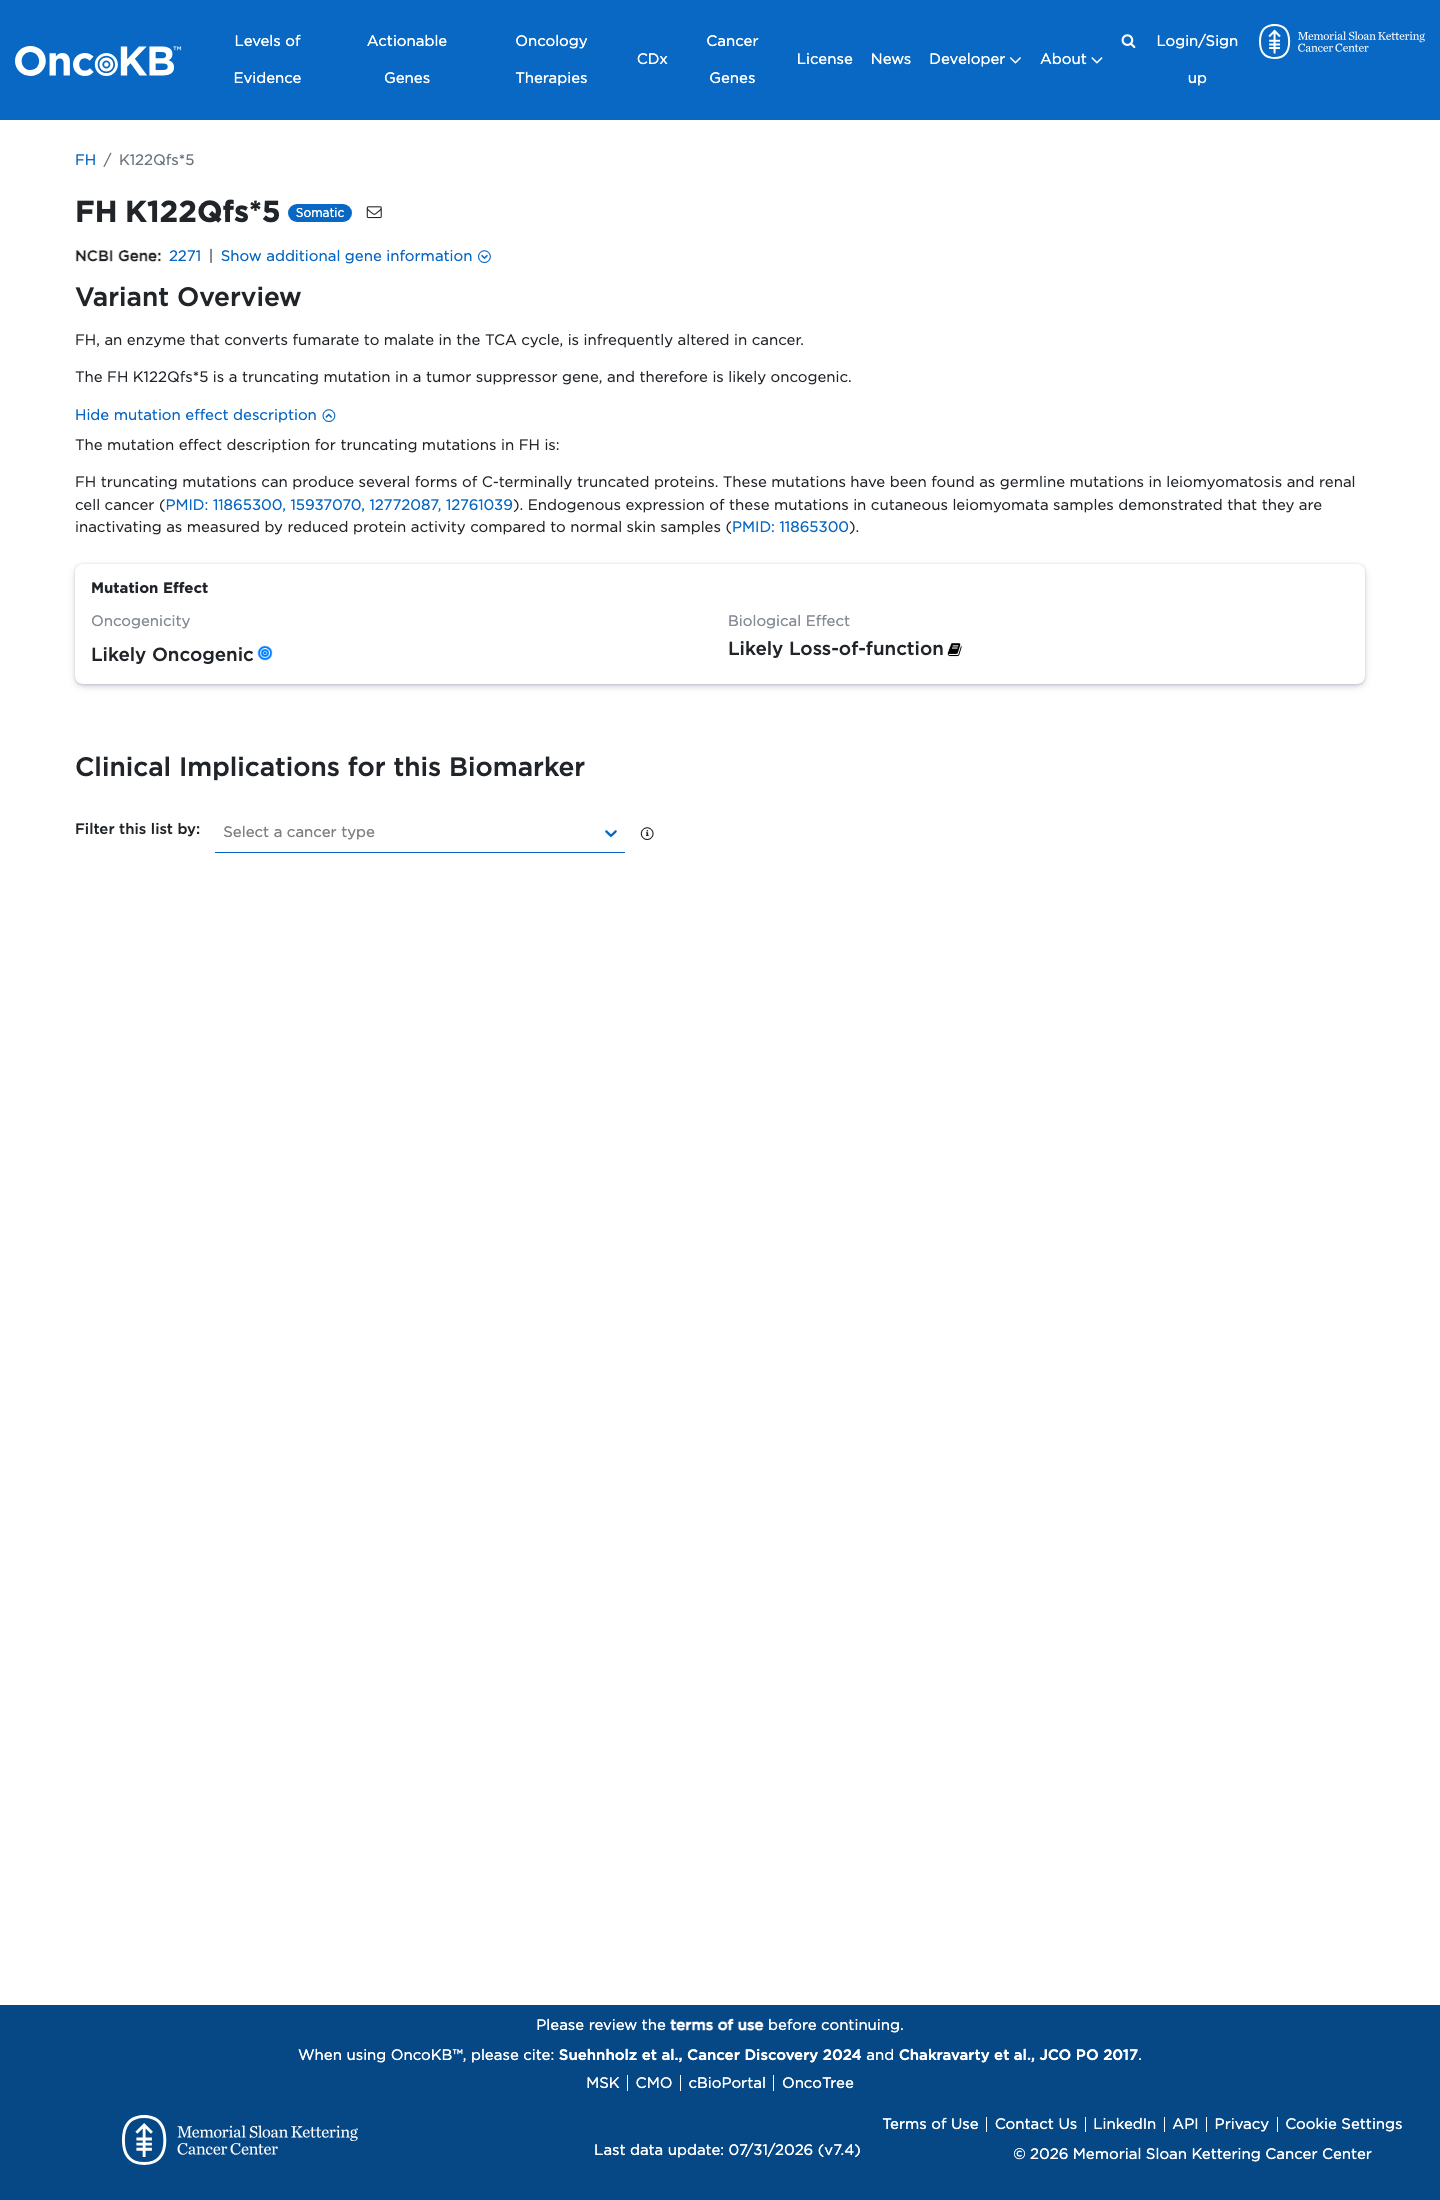
Likely Oncogenic
OncoKB identified variant-specific curated literature and context relevant to functional review; biological-effect context: Likely Loss-of-function; curated oncogenicity label: Likely Oncogenic.
COSMIC
Cancer hotspots
Somatic evidence
Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. COSMIC could not be reviewed because the browser session was redirected to the login page before search results were available.
Hotspots
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
2papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 2 further PMIDs triaged but not cited — see Sources & references.
Rule & framework references · cited for criterion definitions, not variant evidence
11865300 ↗
Germline mutations in FH predispose to dominantly inherited uterine fibroids, skin leiomyomata and papillary renal cell cancer.
12761039 ↗
Genetic and functional analyses of FH mutations in multiple cutaneous and uterine leiomyomatosis, hereditary leiomyomatosis and renal cancer, and fumarate hydratase deficiency.
Sources & reference links
Triaged references · 2 PMIDs not cited in assessment
12772087 ↗
Mutations in the fumarate hydratase gene cause hereditary leiomyomatosis and renal cell cancer in families in North America.
ONCOKB
15937070 ↗
Novel mutations in FH and expansion of the spectrum of phenotypes expressed in families with hereditary leiomyomatosis and renal cell cancer.
ONCOKB