PS1
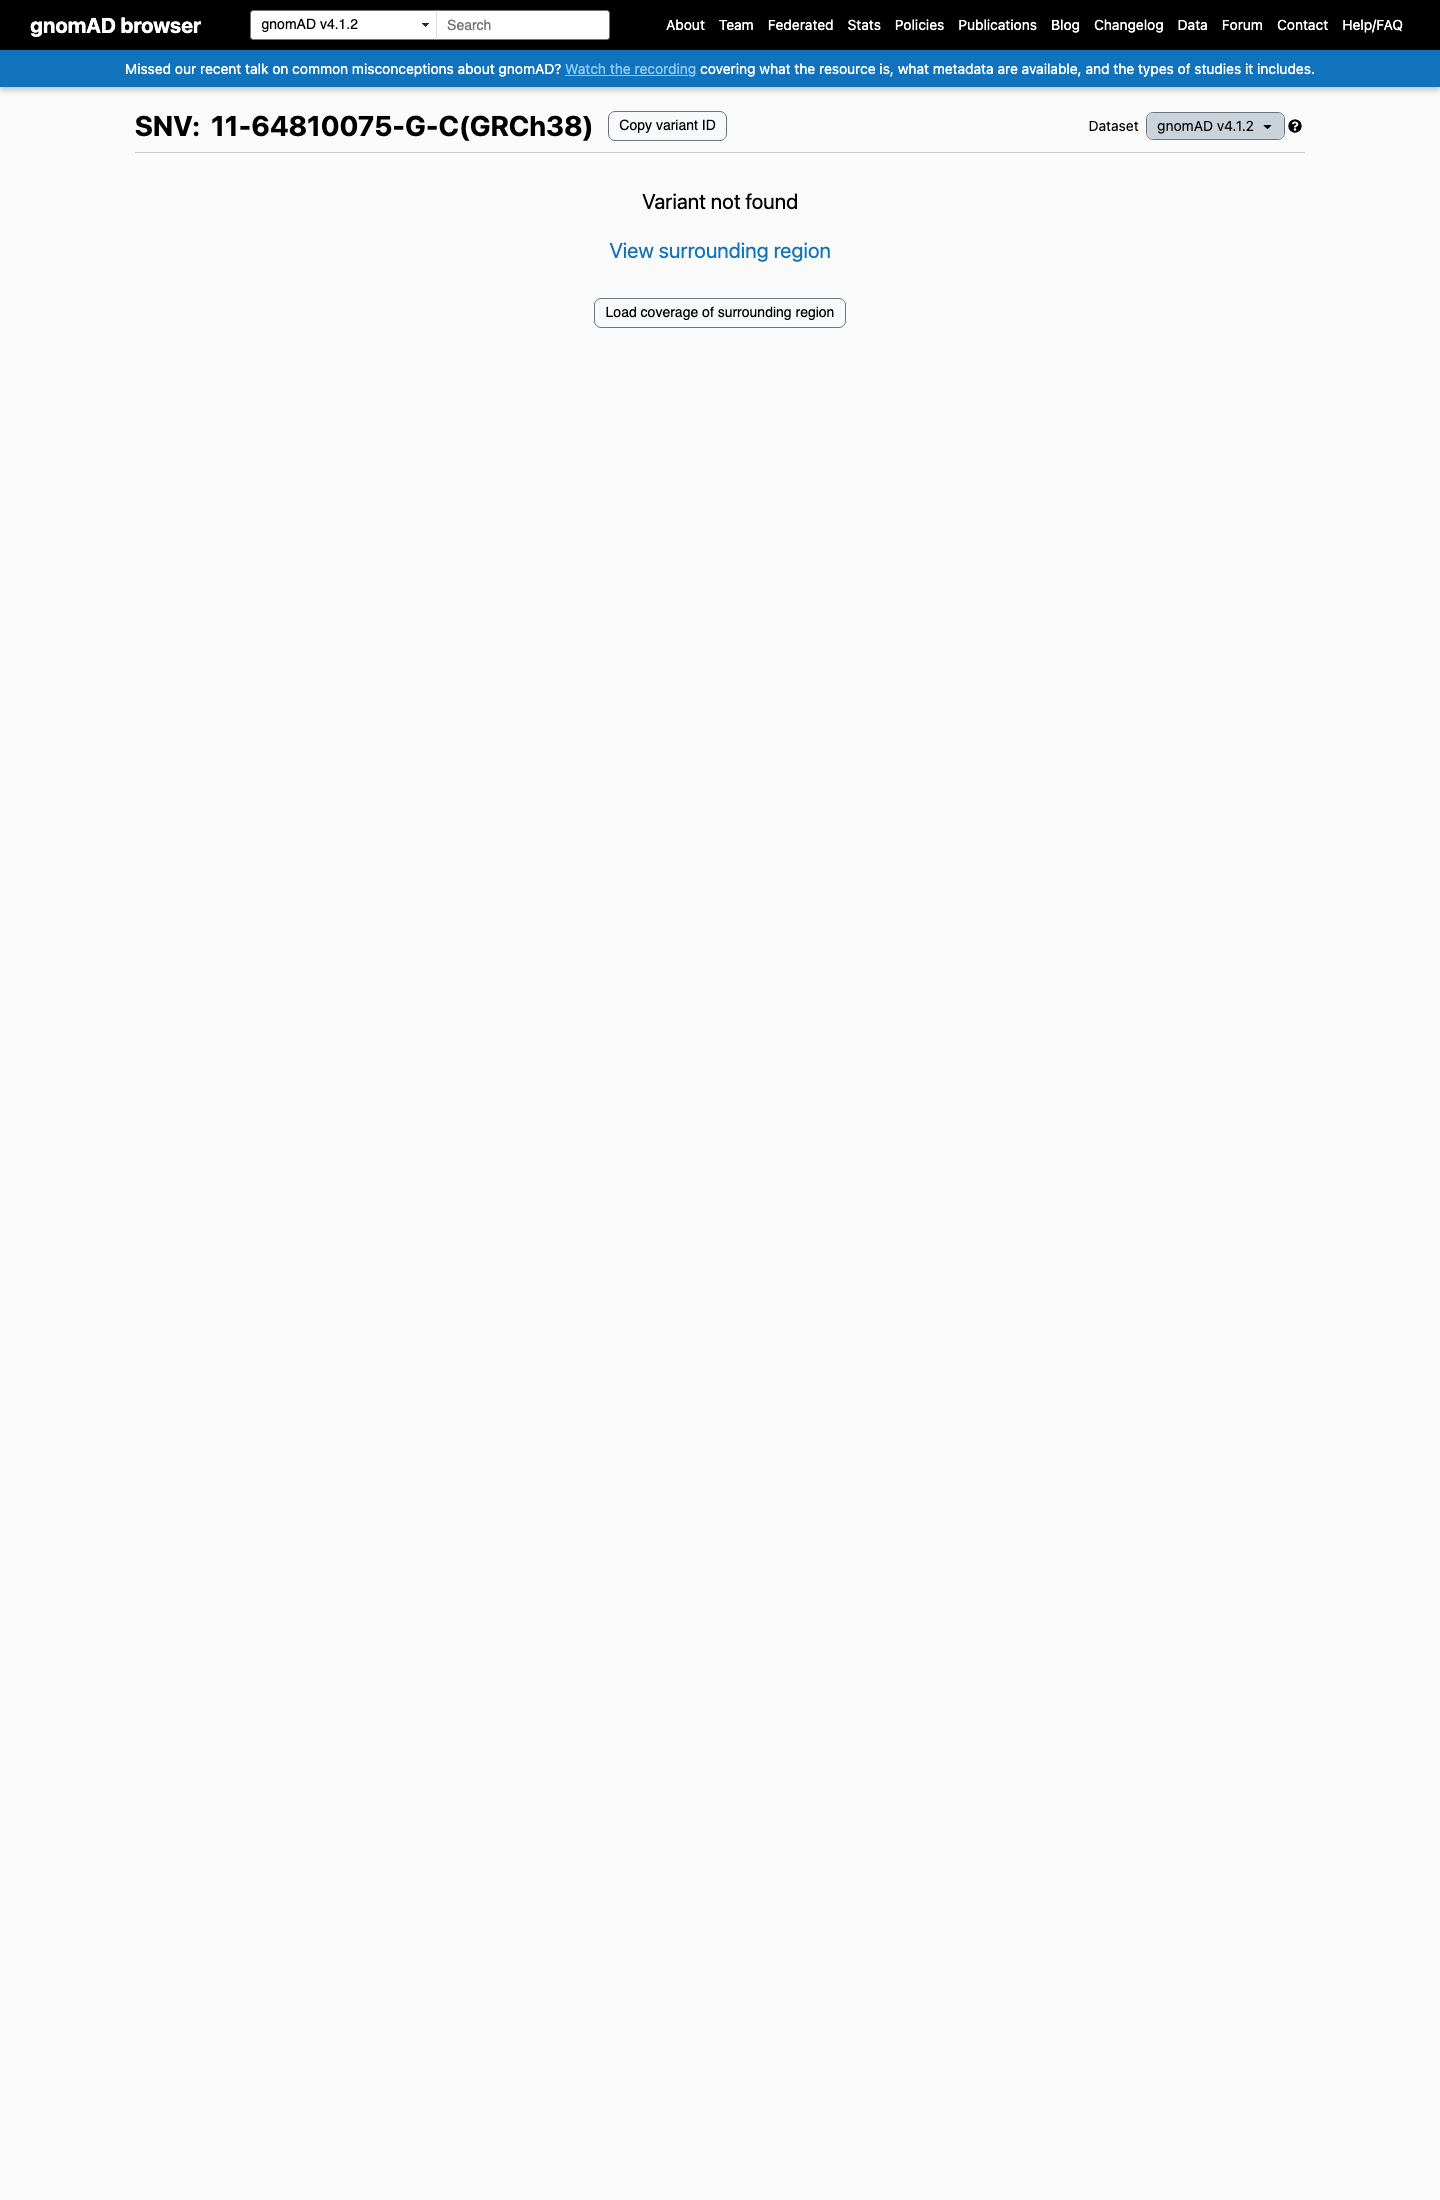
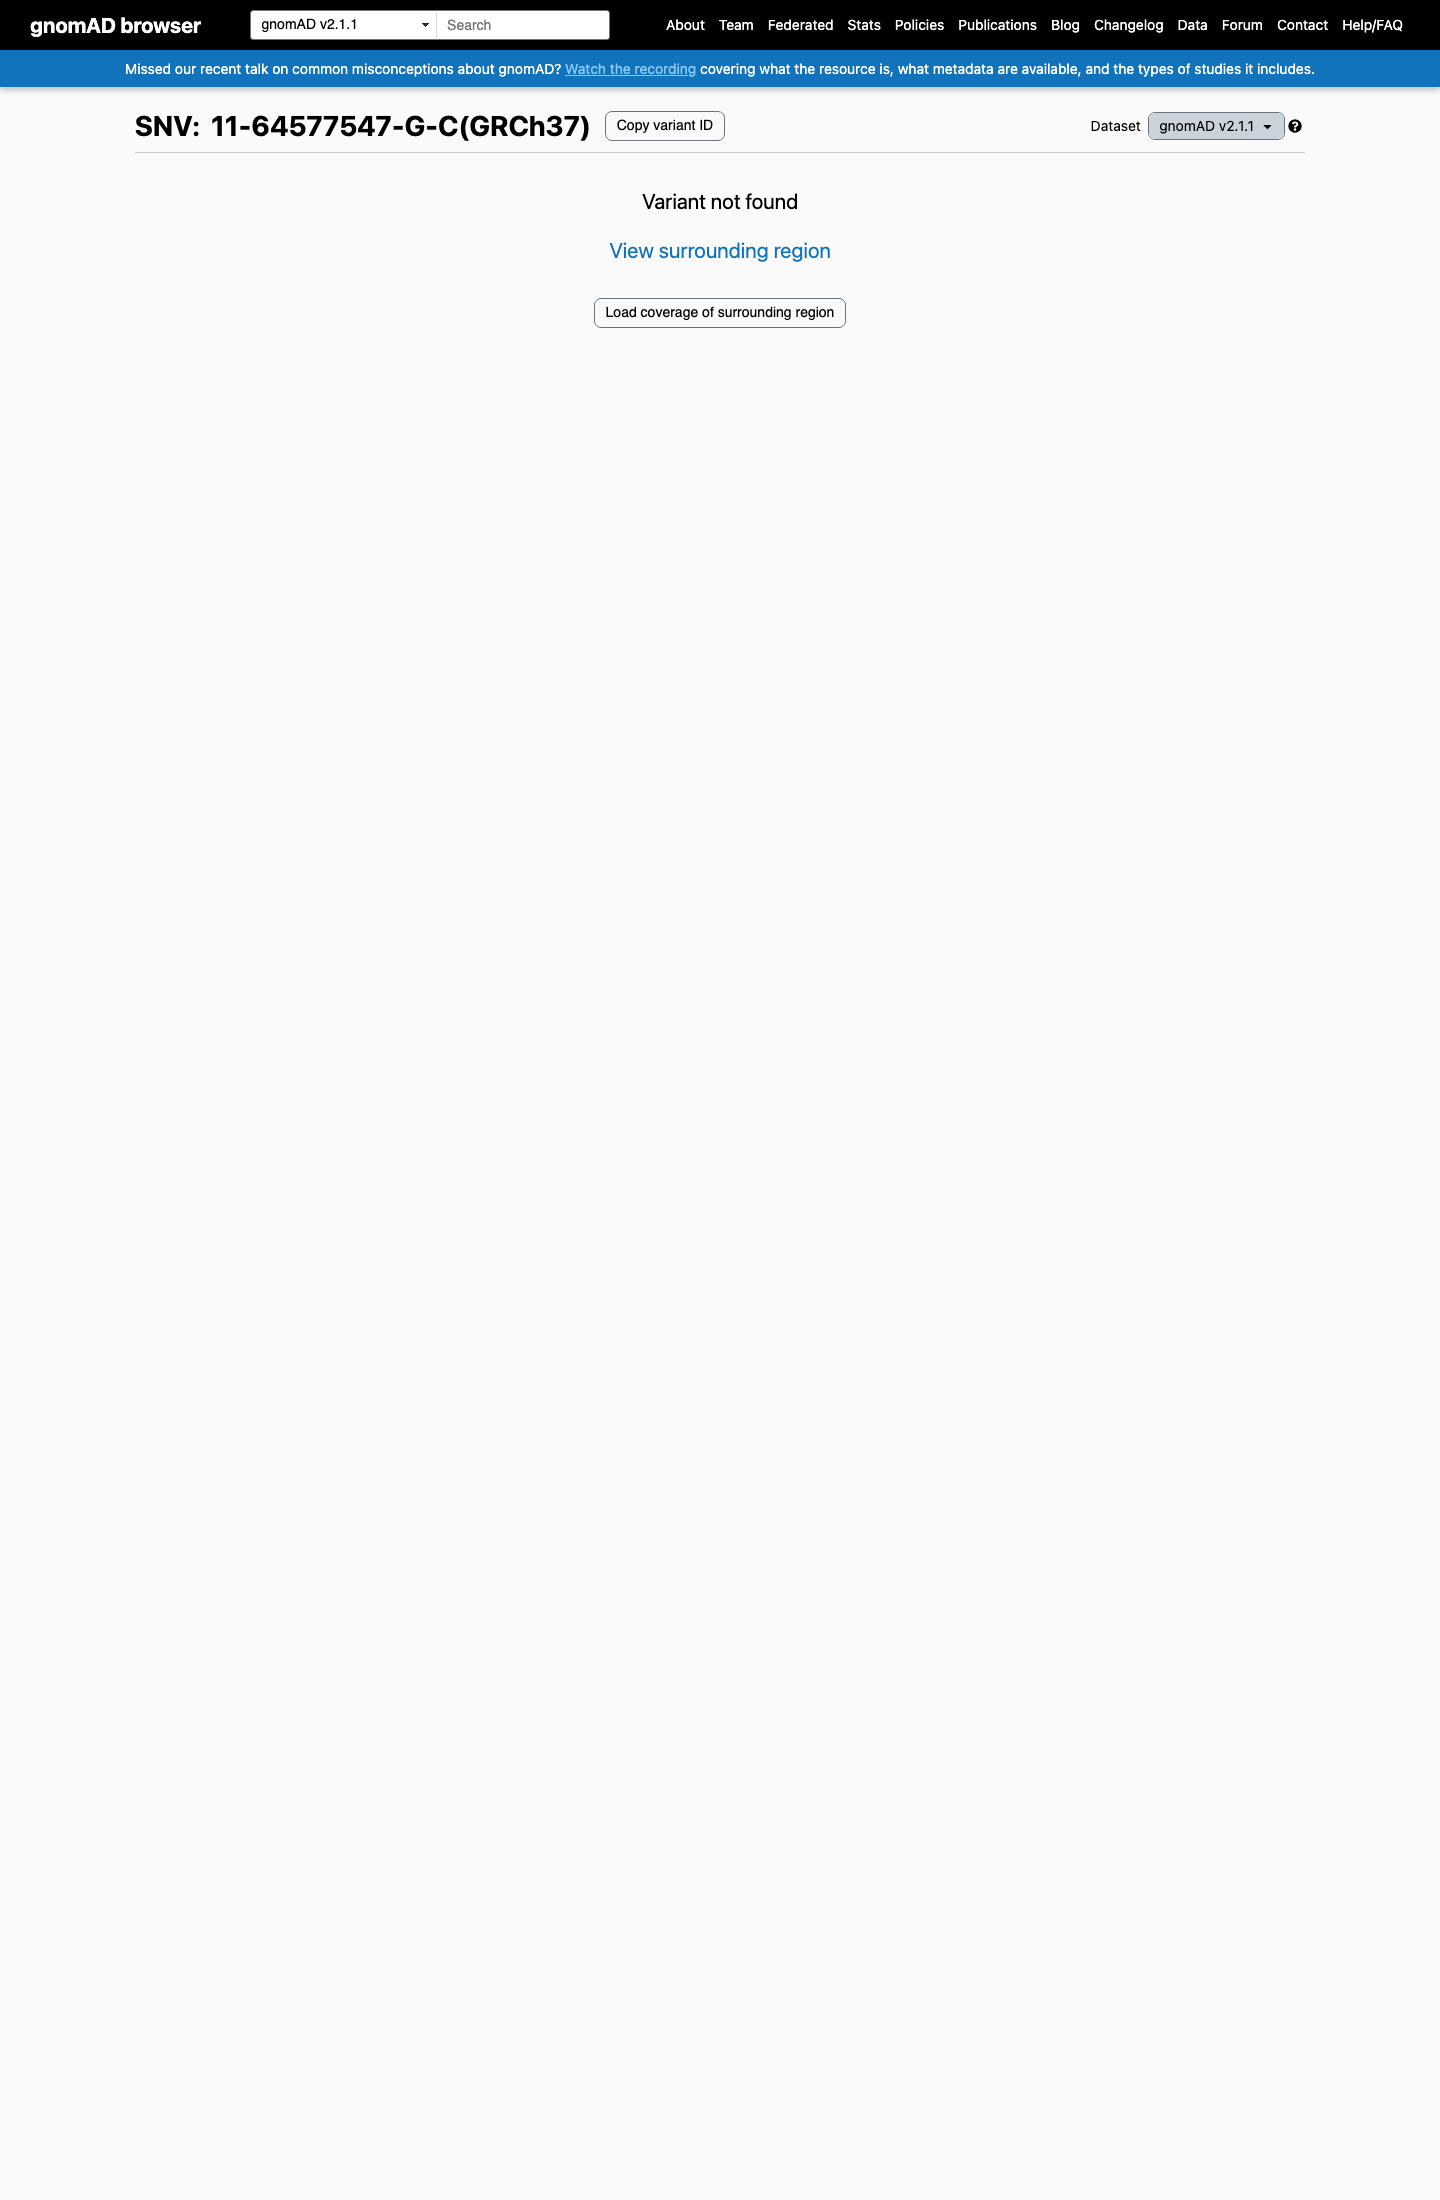
Not assessed: no alternate nucleotide change producing p.Pro12Arg was documented in the available evidence.
PS2
Not assessed: no parental genotypes or confirmed de novo observation are documented for the proband's NM_000244.3:c.35C>G variant.
PS3
Not assessed: no reviewed functional study directly tested MEN1 p.Pro12Arg, while same-codon p.Pro12Leu results cannot establish PS3 for p.Pro12Arg.
PS4
Not assessed: no exact-variant case-control counts or statistically significant phenotype-enrichment metric is available.
PM1
Not assessed: no authoritative MEN1 critical-domain boundary or significant hotspot was available for residue 12.
PM6
Not assessed: no documented phenotype and negative family history support an assumed de novo MEN1 variant in the absence of parental testing.
PP1
Not assessed: no affected relatives, variant-positive family members, or informative meioses are documented for segregation analysis.
PP2
Not met: missense variants comprise approximately 10% of reported MEN1 mutations, so missense is not the predominant disease mechanism.
PP4
Not assessed: no patient-specific, highly specific MEN1 phenotype or family history is documented.
PP5
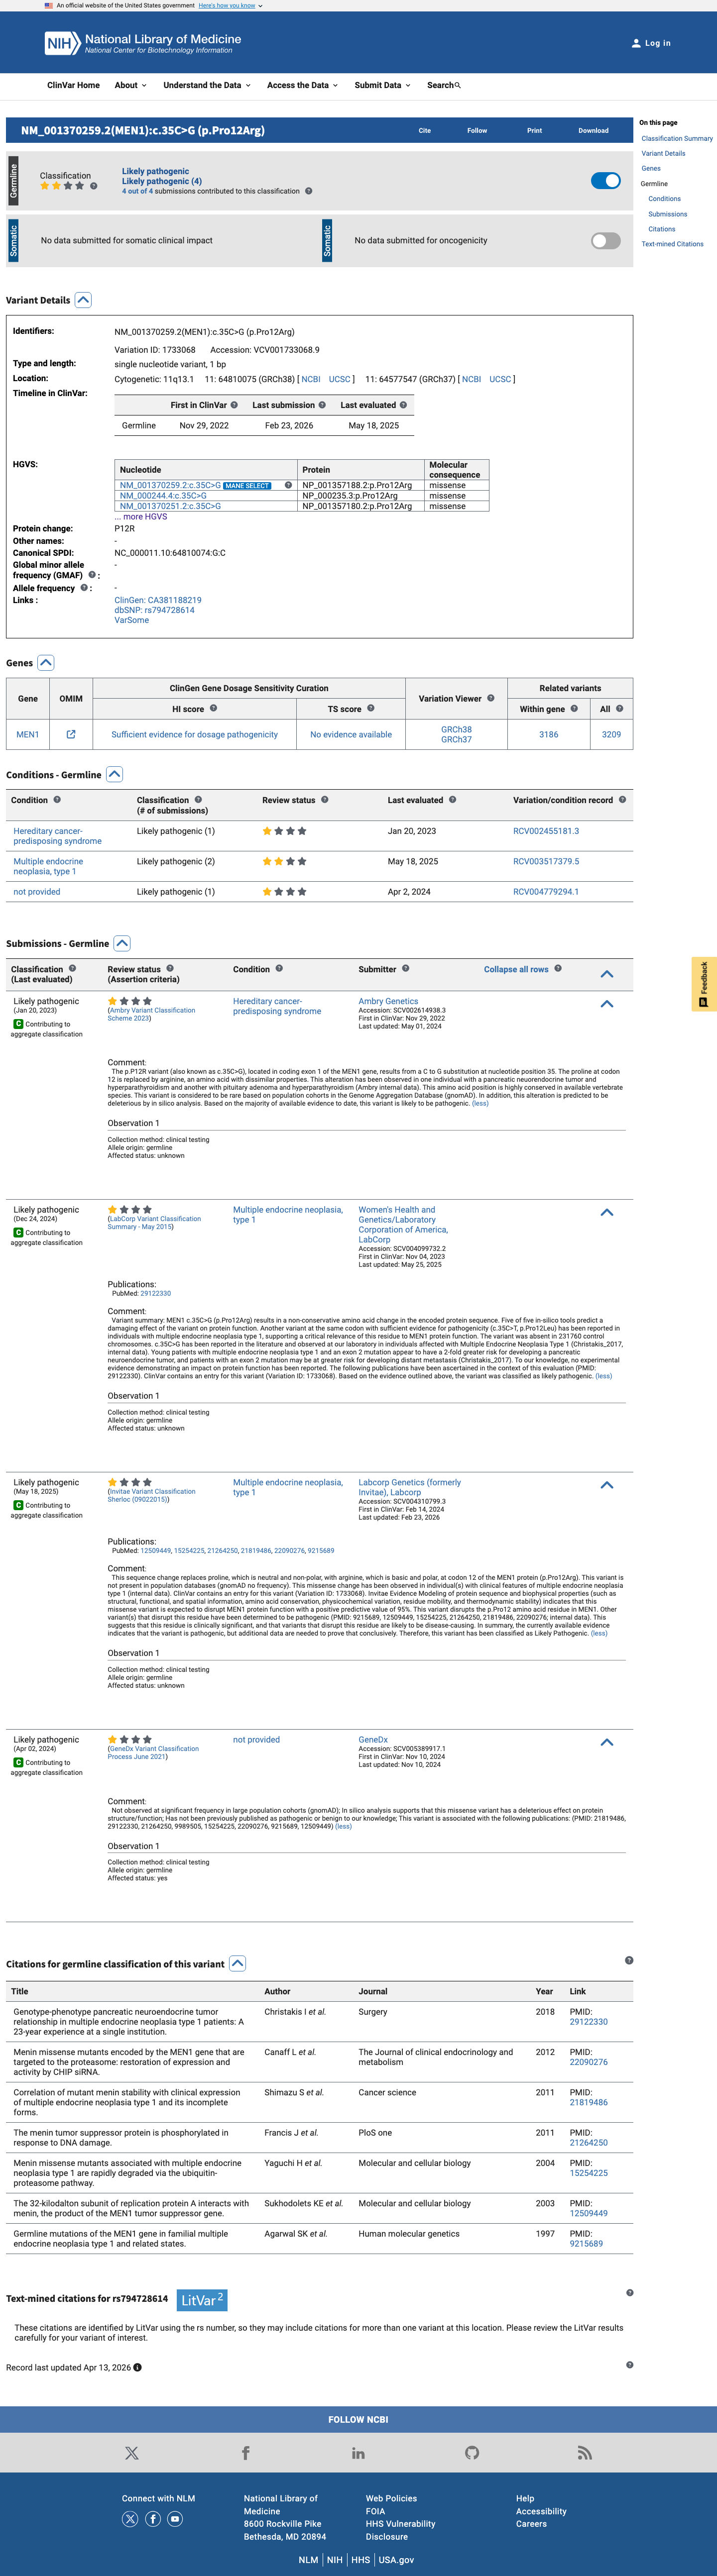
Not met: zero ClinVar expert-panel submissions exist for this exact variant, despite four Likely pathogenic laboratory records.