Back
NM_000465.4:c.1285G>A
p.Glu429Lys · BARD1
ACMG/AMP
0%
complete
Final classification
VUS
BARD1
c.1285G>A
p.Glu429Lys
missense · exon 4

BARD1 encodes a protein that partners with BRCA1 to form a complex essential for repairing damaged DNA, particularly double-stranded breaks, and for maintaining genome stability through cell-cycle checkpoints and chromatin regulation. Germline mutations in BARD1 predispose to breast and ovarian cancer, among other cancers, and the gene is generally regarded as a tumor suppressor, although some evidence suggests it can also promote cancer in certain cellular contexts.

This variant

BARD1 partners with BRCA1 to repair DNA double-strand breaks, and germline mutations in the gene predispose to breast and ovarian cancer. Classifying p.Glu429Lys as a variant of uncertain significance means there is currently no evidence that this specific missense change disrupts BARD1's DNA-repair function or alters cancer risk. Functional or familial data will be needed to determine whether this variant is clinically meaningful.

Transcript
NM_000465.4
HGVS · transcript:coding
NM_000465.4:c.1285G>A
GRCh38
chr2:214780589 C>T
GRCh37
chr2:215645313 C>T
Variant of Uncertain Significance: no ACMG/AMP criterion was met, so no pathogenic or benign combination threshold was reached under the generic ACMG/AMP 2015 framework. Flagged for human review: criteria PM2, BA1, BS1, and BS2 remain unevaluated pending manual confirmation.
Classification rationale
VUS
BARD1 c.1285G>A missense · exon 4

No ACMG/AMP criterion was met, so no pathogenic or benign combination threshold was satisfied; under the generic ACMG/AMP 2015 fallback this resolves to Variant of Uncertain Significance.

LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_000465.4 · variants mapped to exon structure
BARD1 NM_000465.4
Fetching transcript structure from UCSC…
Applied criteria · 0 applied · 23 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 0

No criteria were applied for this variant.

Assessed · not applied · 7 not met · 16 not assessed
Pathogenic
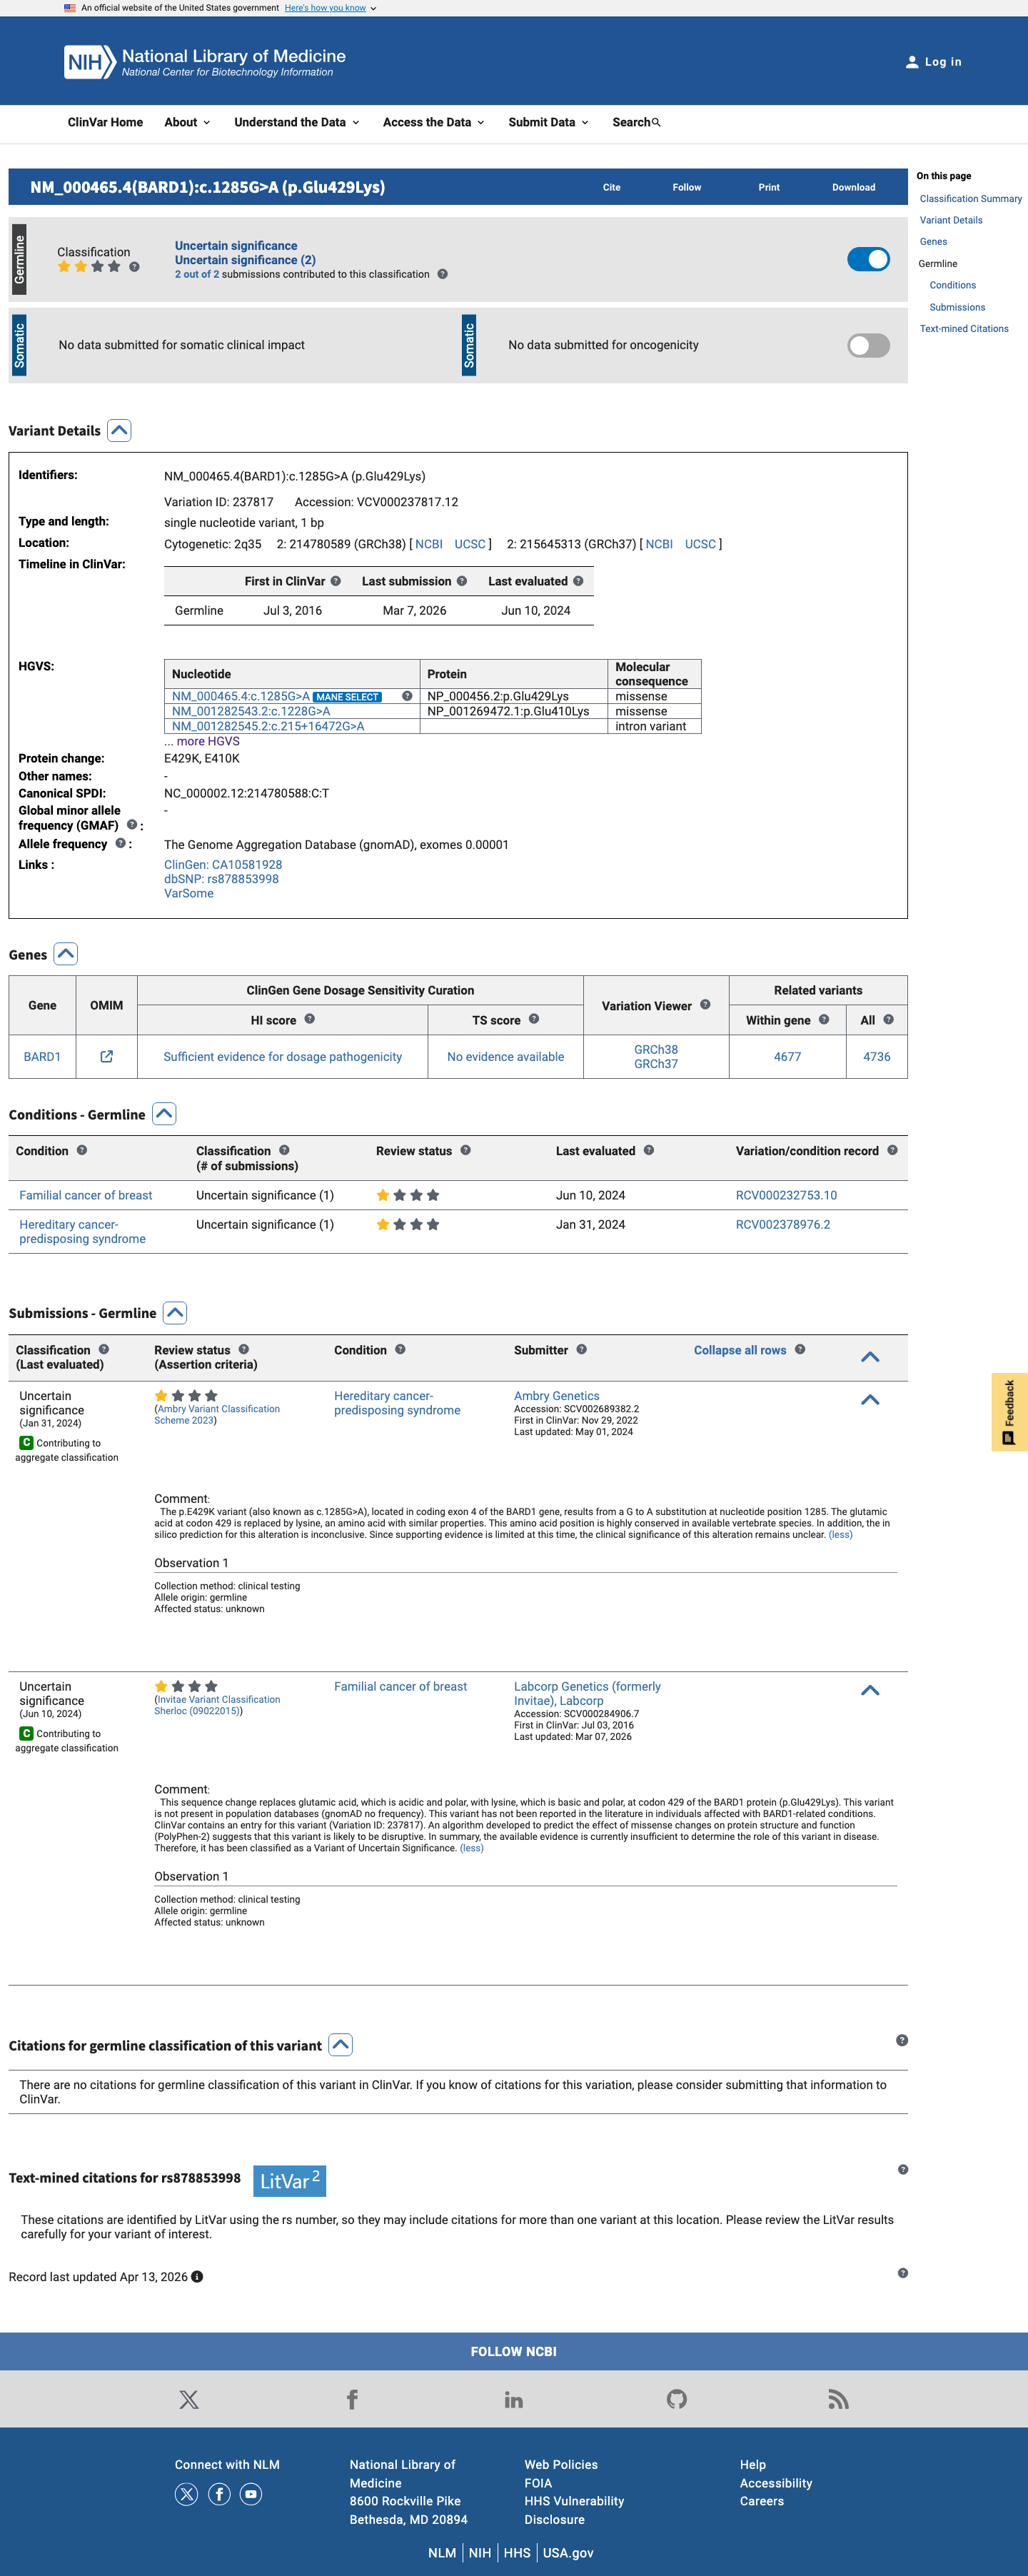
PS1 Not met: the only ClinVar classifications of p.Glu429Lys are two laboratory submissions of uncertain significance, with no pathogenic assertion for this amino acid change.
PS2 Not assessed: no de novo occurrence of this variant with confirmed parentage has been reported in ClinVar or the literature.
PS3 Not assessed: no variant-specific functional assay data exist for p.Glu429Lys; neither OncoKB nor either ClinVar submission reports any functional study.
PS4 Not assessed: no case-control or cohort enrichment data exist for this exact variant in ClinVar or the reviewed literature.
PM1 Not met: p.Glu429Lys is not in a statistically significant cancer hotspot and has no record in COSMIC somatic data.
PM2 Not assessed: insufficient population-frequency evidence was available to evaluate PM2.
PM5 Not assessed: no alternate pathogenic missense at codon 429 was identified, but the same-residue search was incomplete, so evidence is insufficient.
PM6 Not assessed: no assumed de novo occurrence (parental testing without confirmed parentage) has been documented for this variant.
PP1 Not assessed: no co-segregation data in affected family members were available.
PP2 Not assessed: no missense-constraint metric (such as a gnomAD Z-score) or curated missense-versus-truncating distribution was available for BARD1.
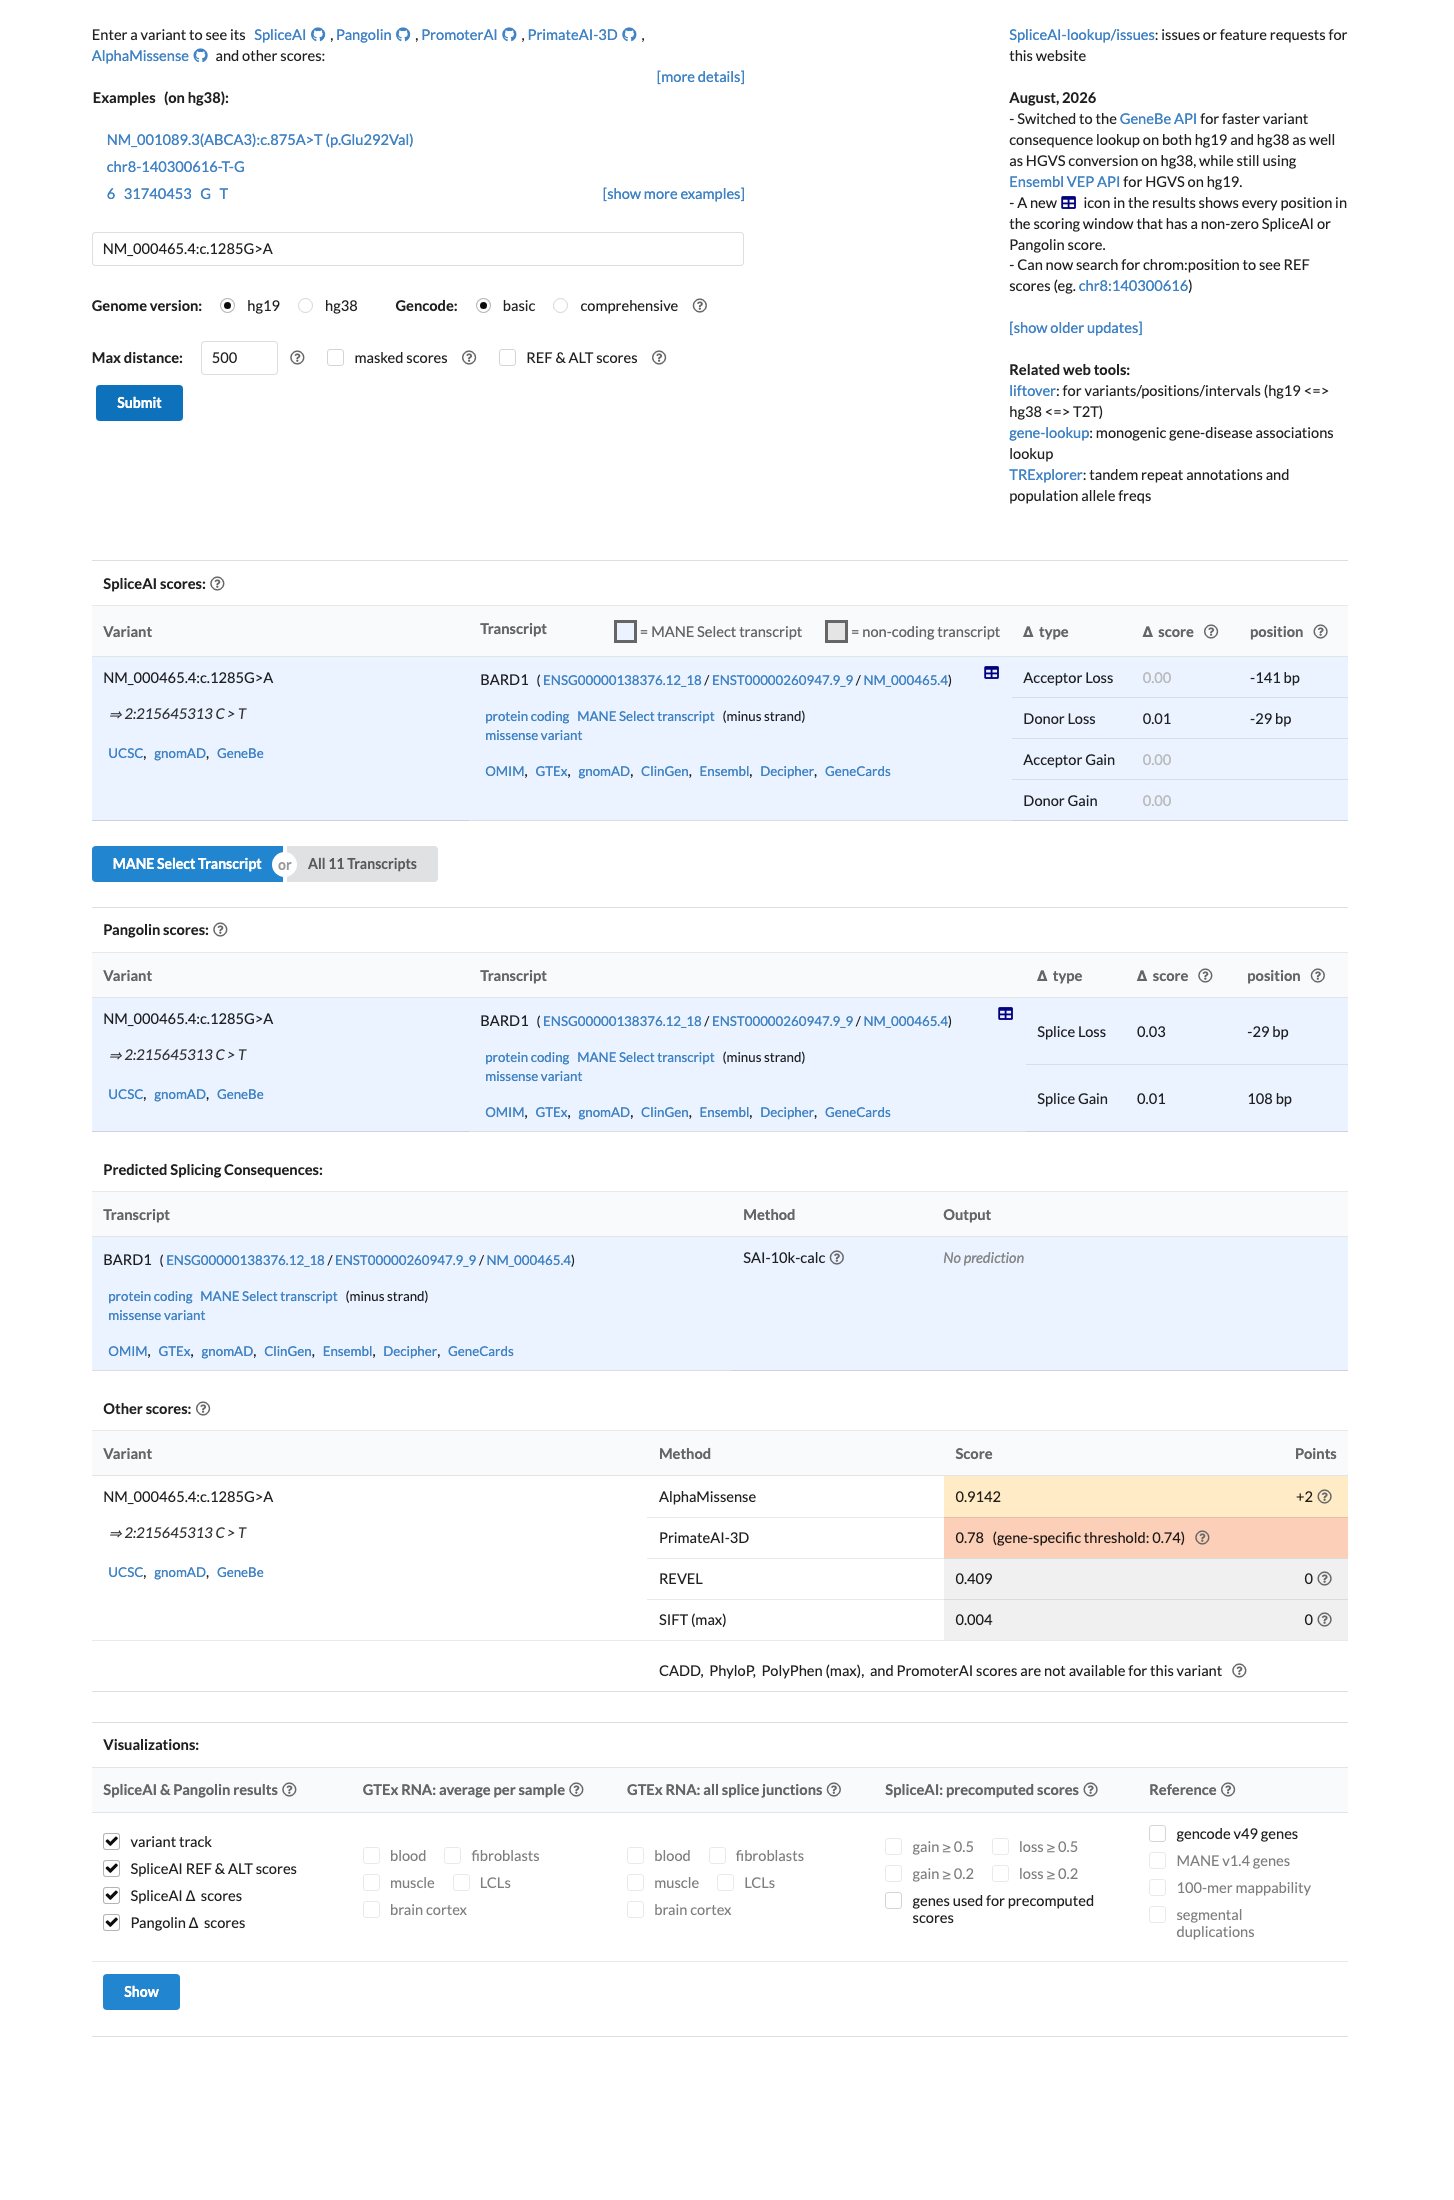
PP3 Not met: REVEL 0.409 falls within the 0.250–0.750 gray zone, below the >0.750 PP3-supporting cutoff.
PP4 Not assessed: no proband-level phenotype or family history was available to show a highly specific single-etiology presentation.
PP5 Not met: no ClinVar expert panel has classified this exact variant as pathogenic or likely pathogenic (expert-panel submissions: 0).
Benign
BA1 Not assessed: insufficient population-frequency evidence was available to evaluate BA1.
BS1 Not assessed: insufficient population-frequency evidence was available to evaluate BS1.
BS2 Not assessed: no observations of this variant in unaffected individuals were available.
BS3 Not assessed: no functional assay data exist for p.Glu429Lys, and the absence of such data cannot itself support a benign effect.
BS4 Not assessed: no family-based non-segregation observations (affected relatives lacking the variant) were available.
BP1 Not assessed: available evidence does not establish that BARD1 disease is primarily caused by truncating variants, as BP1 requires.
BP2 Not met: this variant is never observed in trans with a pathogenic allele, and BARD1 predisposition is not a fully dominant disorder.
BP4 Not met: REVEL 0.409 is above the <0.250 BP4-supporting cutoff, so the variant falls in the gray zone.
BP5 Not assessed: no data show an alternative molecular cause of disease in any carrier of this variant.
BP6 Not met: no ClinVar expert panel has classified this exact variant as benign or likely benign (expert-panel submissions: 0).
N/A · 5 PVS1 · PM3 · PM4 · BP3 · BP7
Research & evidence
Population frequency
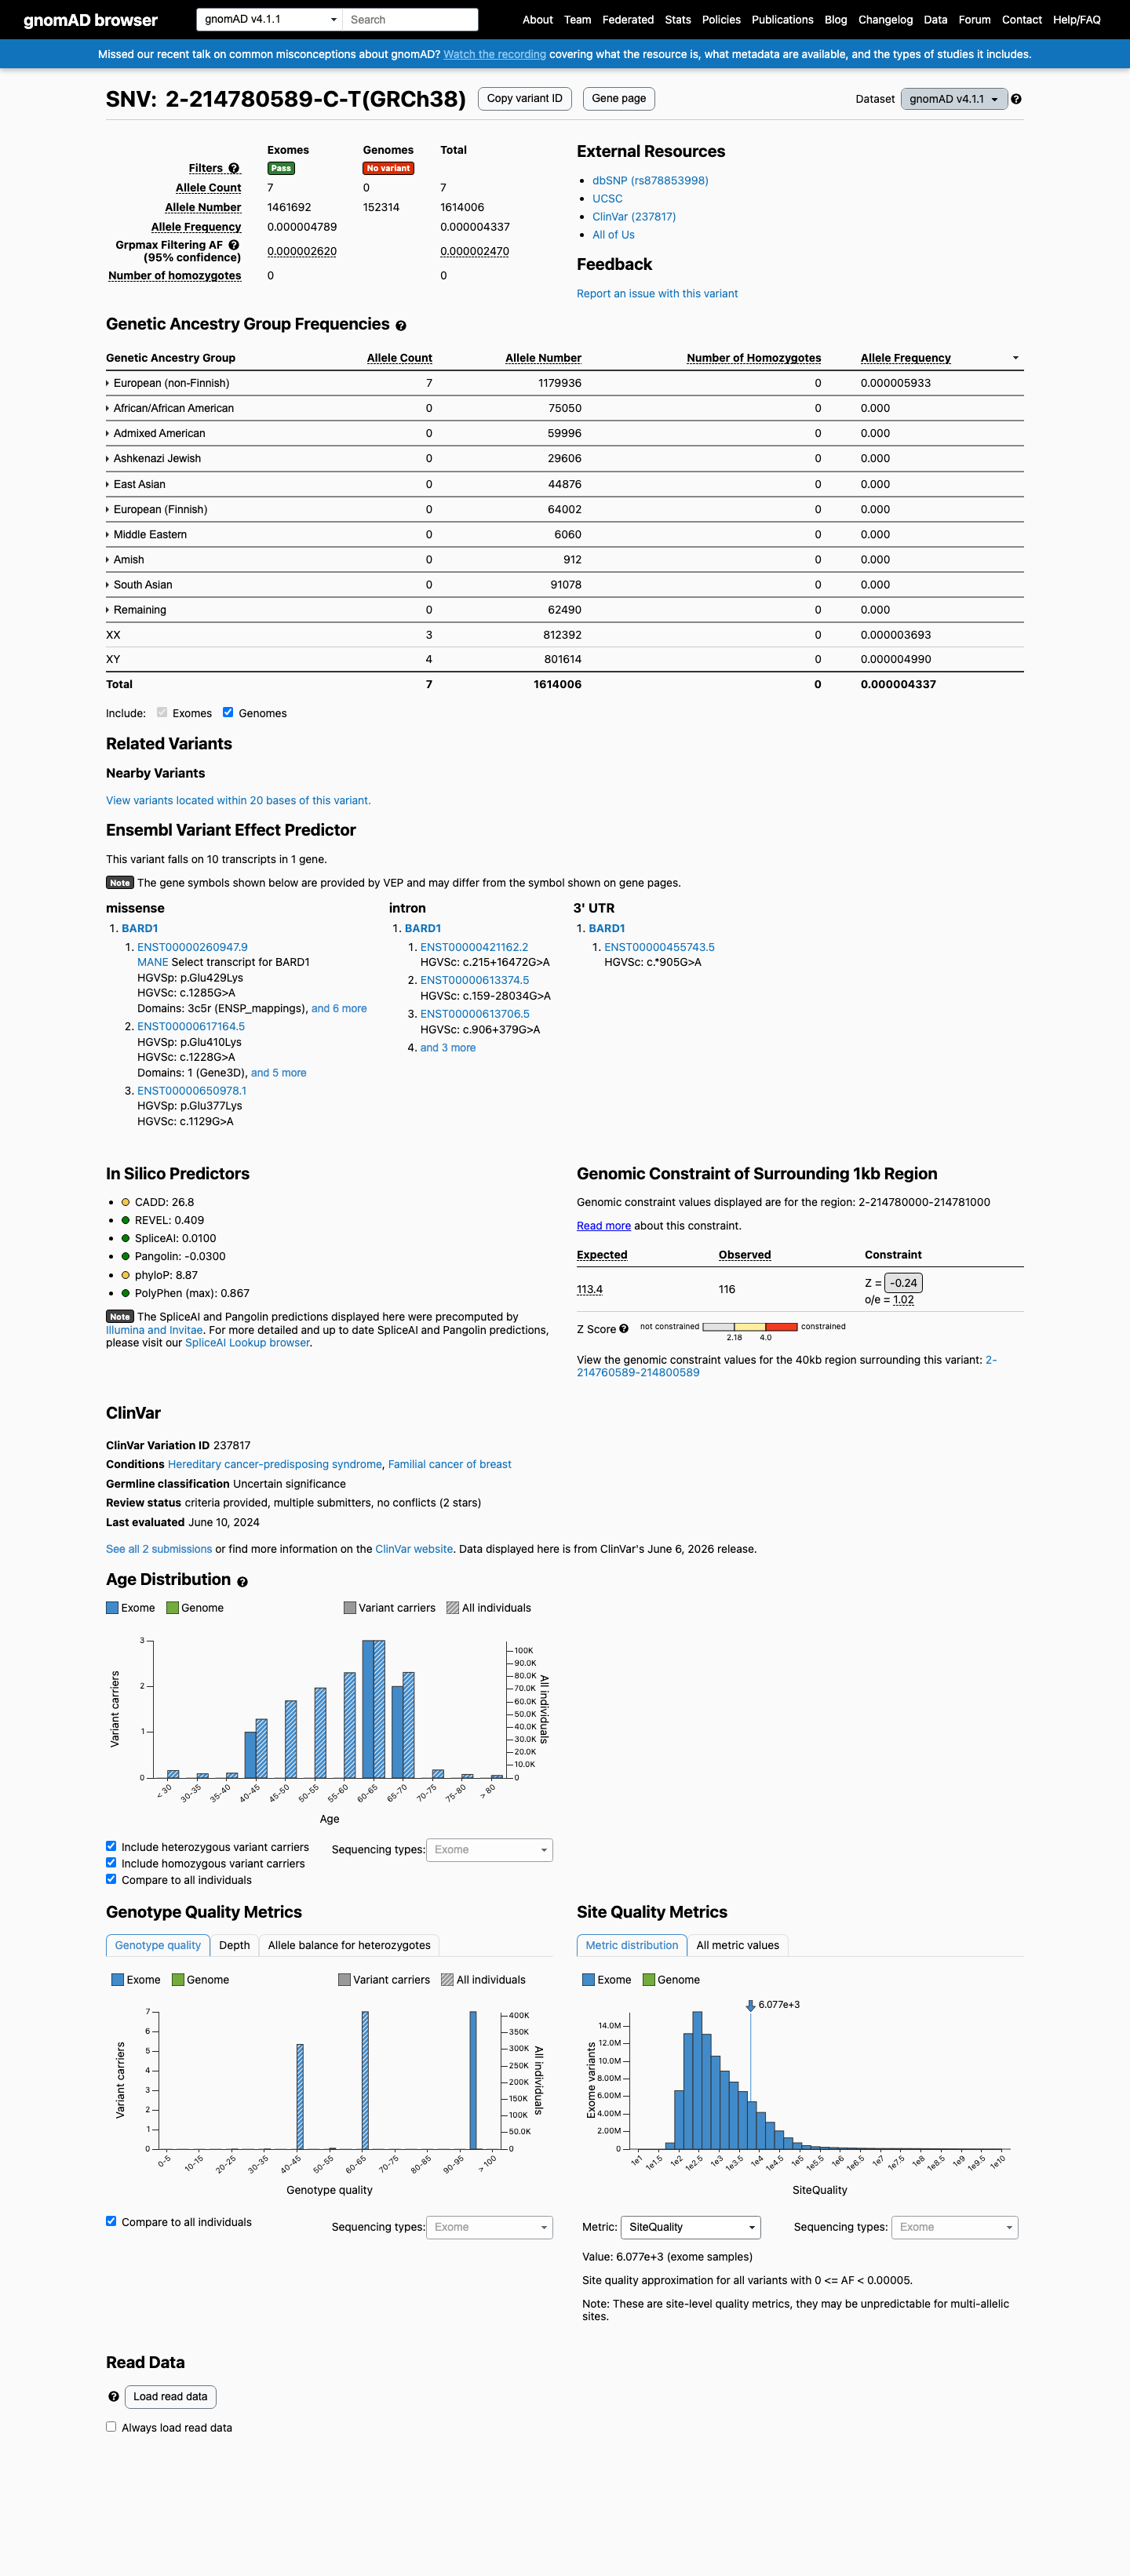
gnomAD v4.1 screenshot
gnomAD v4.1
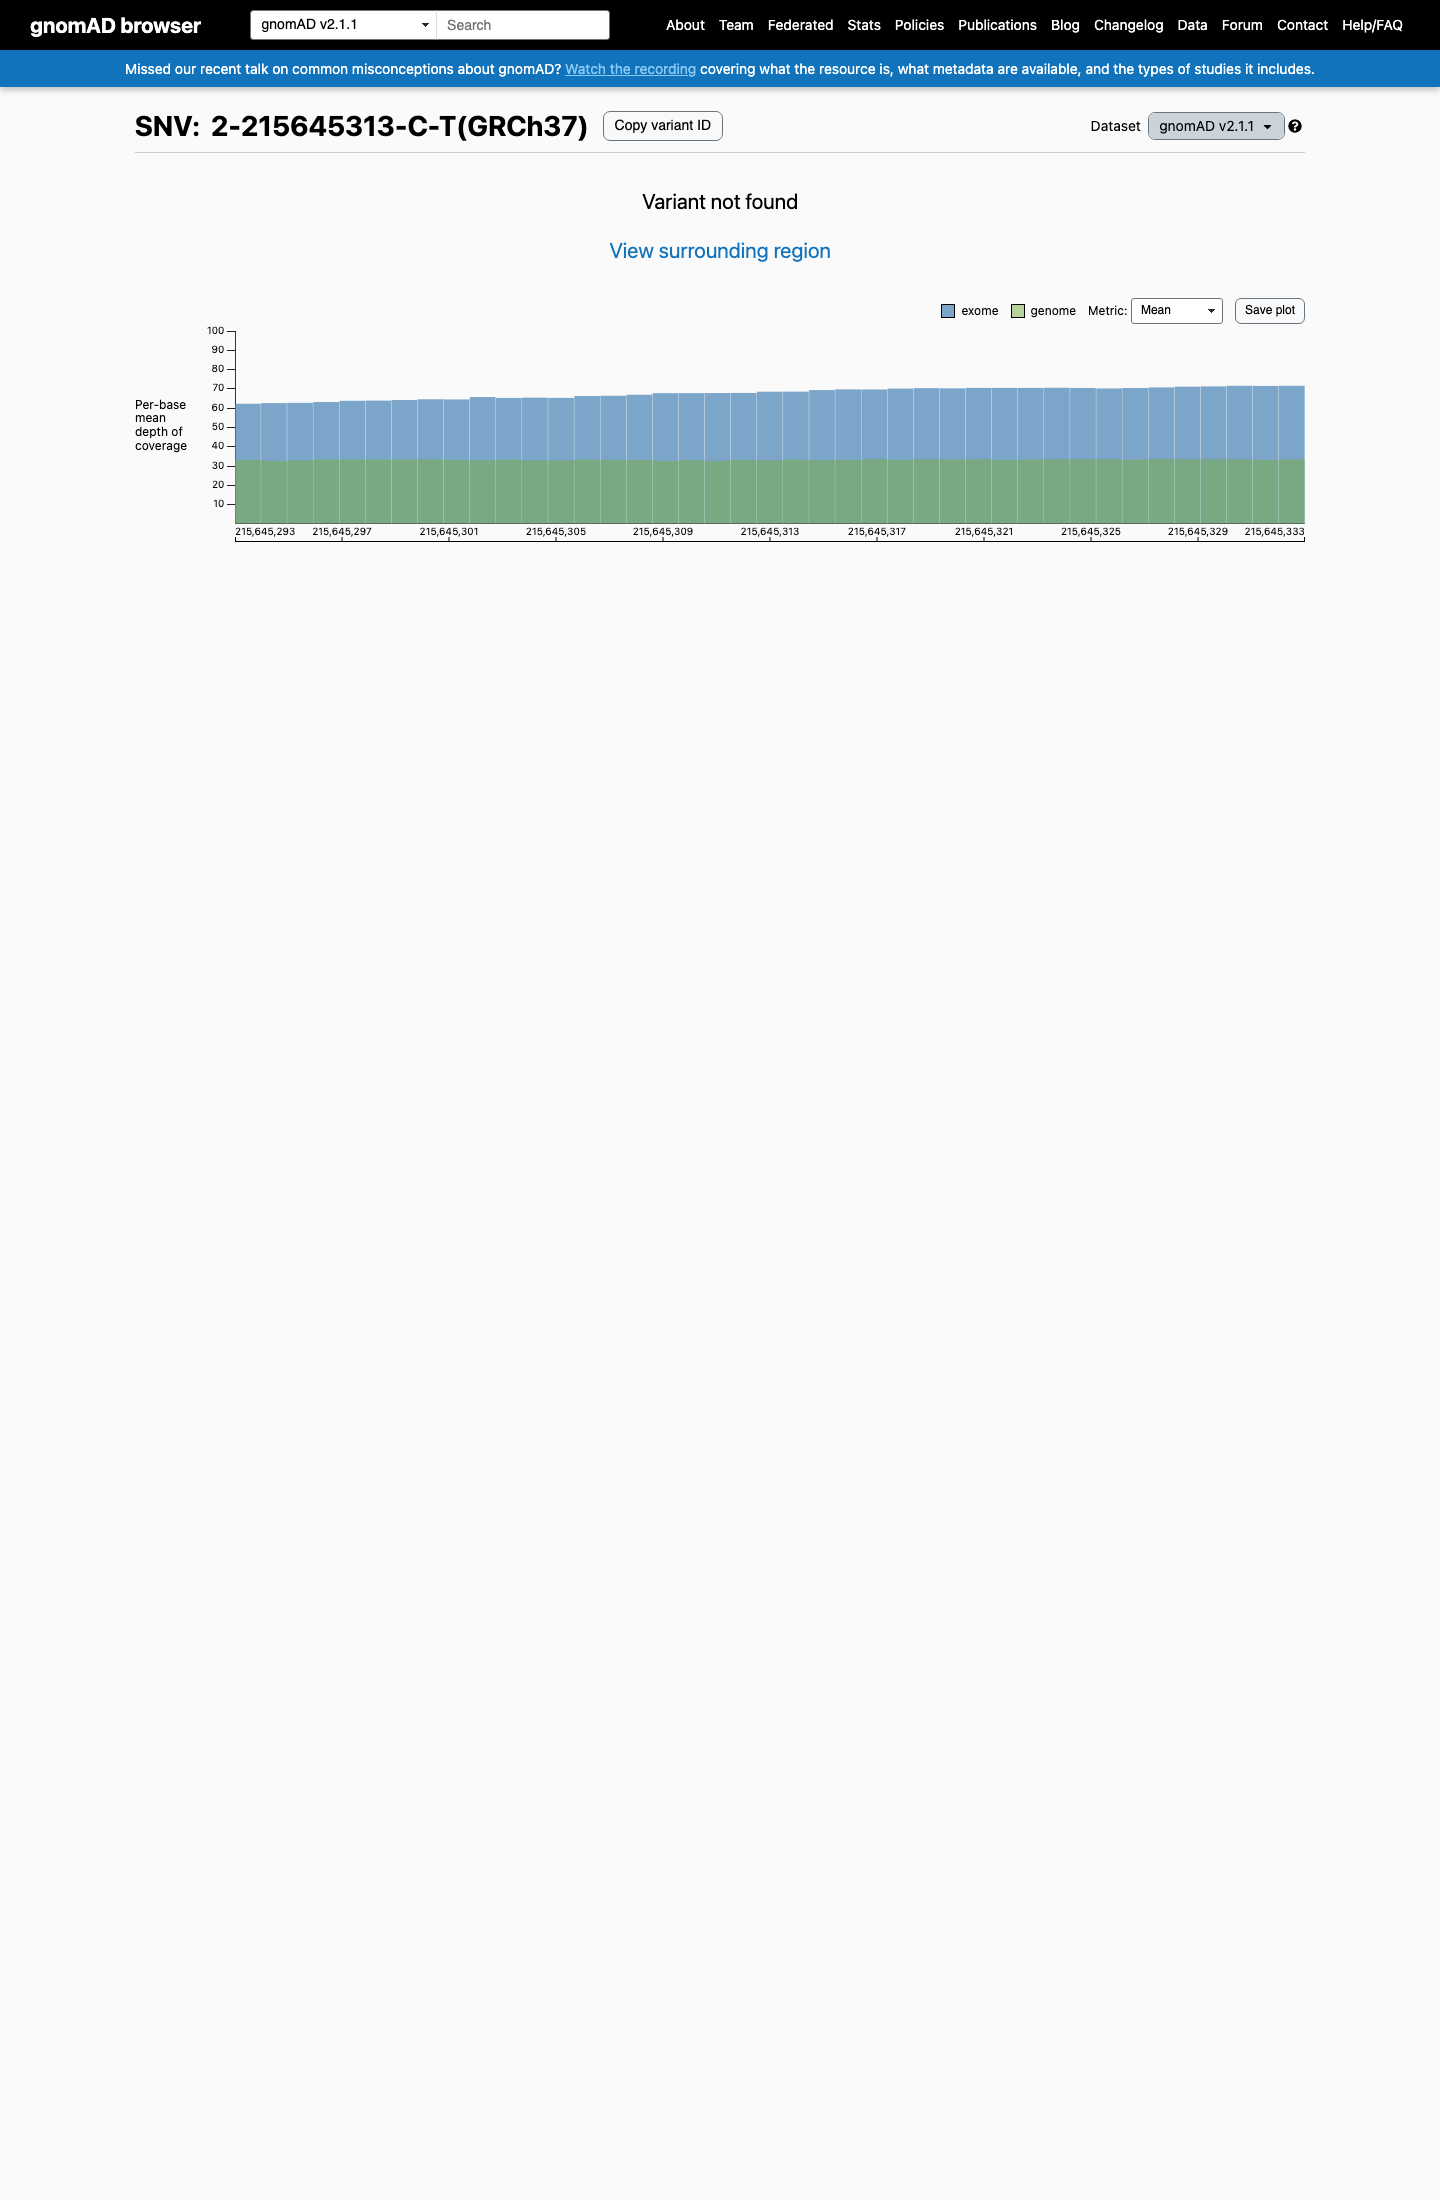
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 4.33703e-06; MAF= 0.00043%, 7/1614006 alleles, homozygotes = 0) and has highest observed frequency in the European (non-Finnish) population (AF= 5.93253e-06; MAF= 0.00059%, 7/1179936 alleles, homozygotes = 0); grpmax FAF= 2.47e-06.
v2.1
Absent from gnomAD v2.1.
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.00043% · 7 / 1,614,006
0 hom · FAF 0.00025%
European (non-Finnish)
7 / 1,179,936
0.00059%
+ 9 not observed (Remaining individuals, Admixed American, European (Finnish), Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American)
gnomAD v2.1
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Uncertain significance (2 clinical laboratories). (ClinVarID = 237817)
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.01). REVEL score = 0.409. BayesDel score = 0.092124.
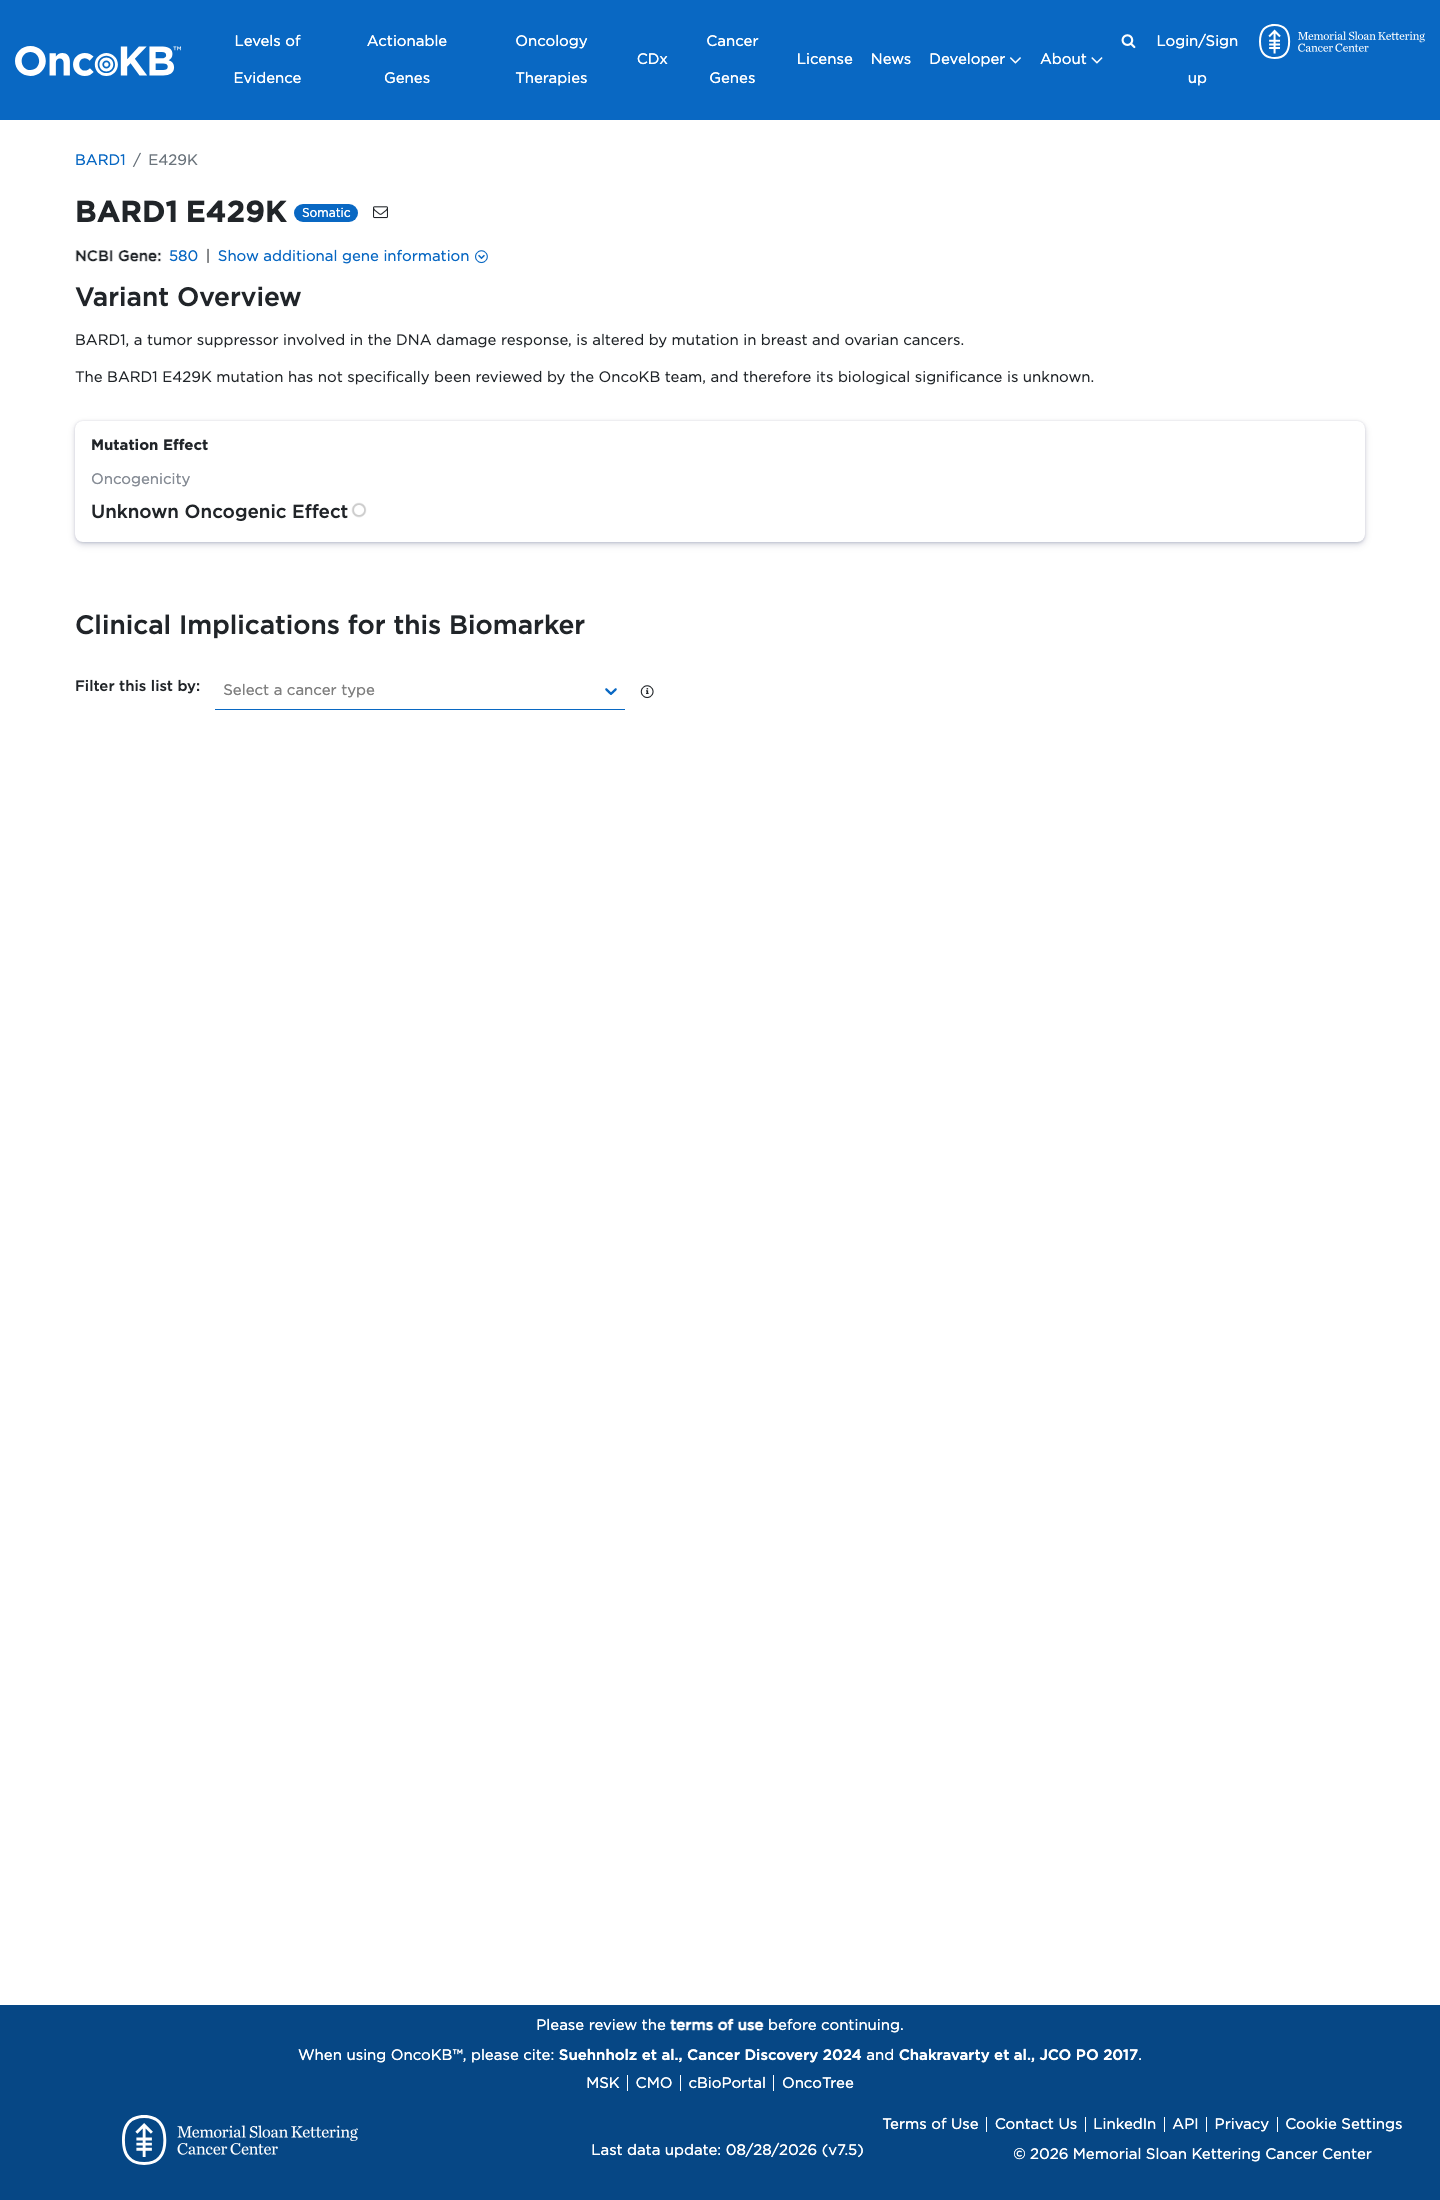
Functional / OncoKB screenshot
Functional Unknown Oncogenic Effect
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. BARD1, a tumor suppressor involved in the DNA damage response, is altered by mutation in breast and ovarian cancers.
OncoKB ↗
COSMIC screenshot
COSMIC
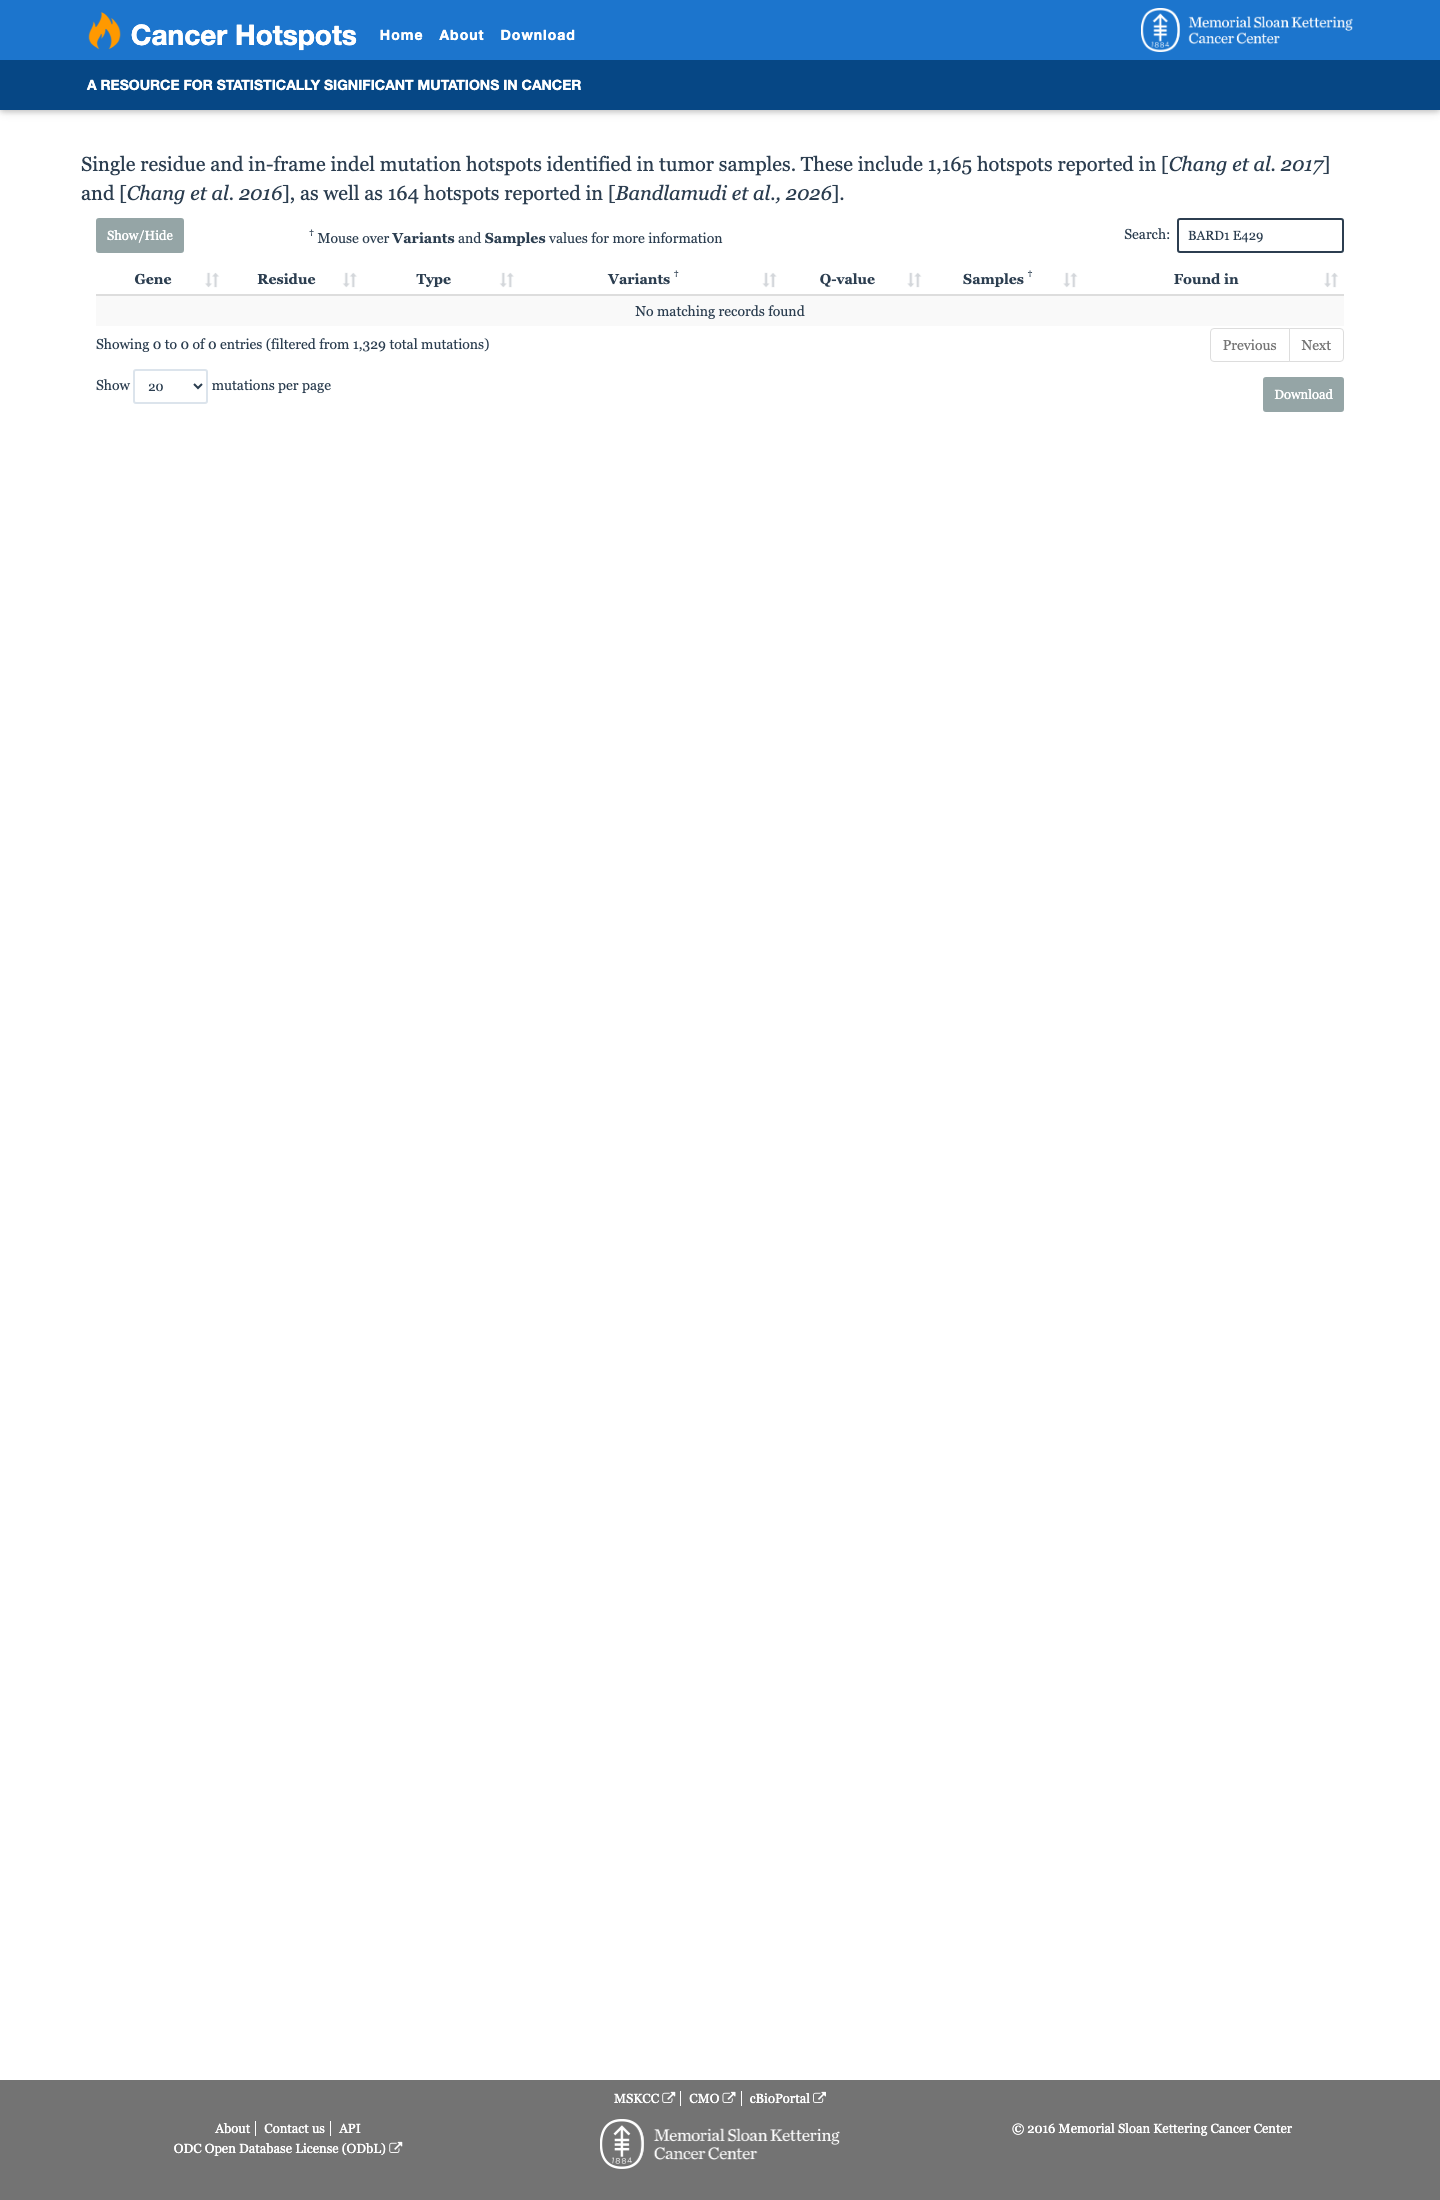
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 5 PMIDs not cited in assessment
17508274 ↗ Risk assessment and genetic counseling for hereditary breast and ovarian cancer: recommendations of the National Society of Genetic Counselors. CLINVAR
26389258 ↗ Cancer Genetics Risk Assessment and Counseling (PDQ®): Health Professional Version. CLINVAR
31429903 ↗ Risk Assessment, Genetic Counseling, and Genetic Testing for BRCA-Related Cancer: US Preventive Services Task Force Recommendation Statement. CLINVAR
31479213 ↗ PMID 31479213 CLINVAR
34012068 ↗ ACMG SF v3.0 list for reporting of secondary findings in clinical exome and genome sequencing: a policy statement of the American College of Medical Genetics and Genomics (ACMG). CLINVAR