Back
NM_000535.7:c.1501G>A
p.Val501Met · PMS2
0%
complete
Final classification
VUS
BP4
PMS2
c.1501G>A
p.Val501Met
missense · exon 11

PMS2 is a DNA mismatch-repair protein that works with MLH1 to correct replication errors, including mismatches and small insertion or deletion loops. It acts as a tumor suppressor, and inherited loss of PMS2 function can cause Lynch syndrome and constitutional mismatch repair deficiency, increasing the risk of colorectal, endometrial, ovarian, urothelial, and other cancers. Loss of PMS2 function in tumors can cause hypermutation and microsatellite instability, which may make some cancers responsive to certain immunotherapies.

This variant

PMS2 encodes a DNA mismatch-repair protein that partners with MLH1, and inherited loss of PMS2 function is associated with Lynch syndrome and constitutional mismatch repair deficiency.

Transcript
NM_000535.7
HGVS · transcript:coding
NM_000535.7:c.1501G>A
GRCh38
chr7:5987264 C>T
GRCh37
chr7:6026895 C>T
VUS: BP4 (supporting) is the only applied criterion, and no PMS2 VCEP Version 2.0 combination rule is satisfied.
Classification rationale
BP4 VUS
PMS2 c.1501G>A missense · exon 11

BP4 Supporting: the PMS2 HCI prior pathogenicity probability is 0.002, below the VCEP BP4 threshold of <0.11.

BP4 VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_000535.7 · variants mapped to exon structure
PMS2 NM_000535.7
Fetching transcript structure from UCSC…
Applied criteria · 1 applied · 15 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 1
Strength Supporting Moderate Strong Very strong
BP4 supporting Benign
Met, Supporting: HCI prior pathogenicity probability 0.002 is below the PMS2 VCEP BP4 threshold of <0.11.
The PMS2 VCEP index declares HCI-PRIORS-PMS2.txt governing for PP3 and BP4 in missense variants.The exact HCI table entry for c.1501G>A / p.V501M assigns pathogenicity probability 0.002.The PMS2 VCEP specification assigns BP4 Supporting to a missense variant with MAPP/PP2 Prior P <0.11; 0.002 satisfies this rule.
Assessed · not applied · 6 not met · 9 not assessed
Pathogenic
PS1 Not met: no alternate-nucleotide PMS2 variant encoding p.Val501Met is established by the VCEP as Pathogenic.
PS2 Not assessed: no proband-specific de novo observation, parental confirmation, LS-spectrum tumor evidence, or de novo point total is documented.
PS3 Not assessed: no variant-specific PMS2 functional assay result or calibrated functional odds are available to meet the VCEP PS3 thresholds of >2.08, >4.3, or >18.7.
PM2 Not met: gnomAD v4.1 allele frequency is 5.6378237256969525e-05, exceeding the PMS2 PM2 threshold of 0.00002.
PM3 Not assessed: no affected-proband observation documents a second pathogenic PMS2 variant in trans or provides phase evidence needed for PM3 points.
PM5 Not met: zero same-residue Val501 comparator candidates were found, and the HCI prior probability is 0.002 versus the >0.68 PP3-supporting threshold.
PP1 Not assessed: no tested relatives, pedigree segregation data, meioses, or combined Bayes likelihood ratio is documented.
PP3 Not met: HCI prior pathogenicity probability 0.002 is below the PMS2 VCEP PP3 supporting threshold of >0.68.
PP4 Not assessed: no variant-specific MSI, tumor-genome, or PMS2-consistent IHC result is available to compare with the VCEP's one-tumor PP4 threshold.
Benign
BA1 Not met: gnomAD v4.1 grpmax filtering allele frequency is 5.628e-05, below the PMS2 BA1 threshold of 0.0028.
BS1 Not met: gnomAD v4.1 grpmax filtering allele frequency is 5.628e-05, below the PMS2 BS1 lower threshold of 0.00028.
BS2 Not assessed: gnomAD v4.1 shows zero homozygotes, but the PMS2 BS2 rule requires documented in-trans clinical co-occurrence and confirmed phase.
BS3 Not assessed: no variant-specific proficient assay result or calibrated functional odds are available to meet the VCEP BS3 thresholds of <=0.05 or >0.05 and <=0.48.
BS4 Not assessed: no non-segregating affected relatives, pedigree analysis, or combined Bayes likelihood ratio is documented.
BP5 Not assessed: no qualifying tumor phenotype, BRAF V600E, or MLH1 methylation result is available for comparison with the BP5 tumor-count thresholds.
N/A · 12 PVS1 · PS4 · PM1 · PM4 · PM6 · PP2 · PP5 · BP1 · BP2 · BP3 · BP6 · BP7
Research & evidence
Population frequency
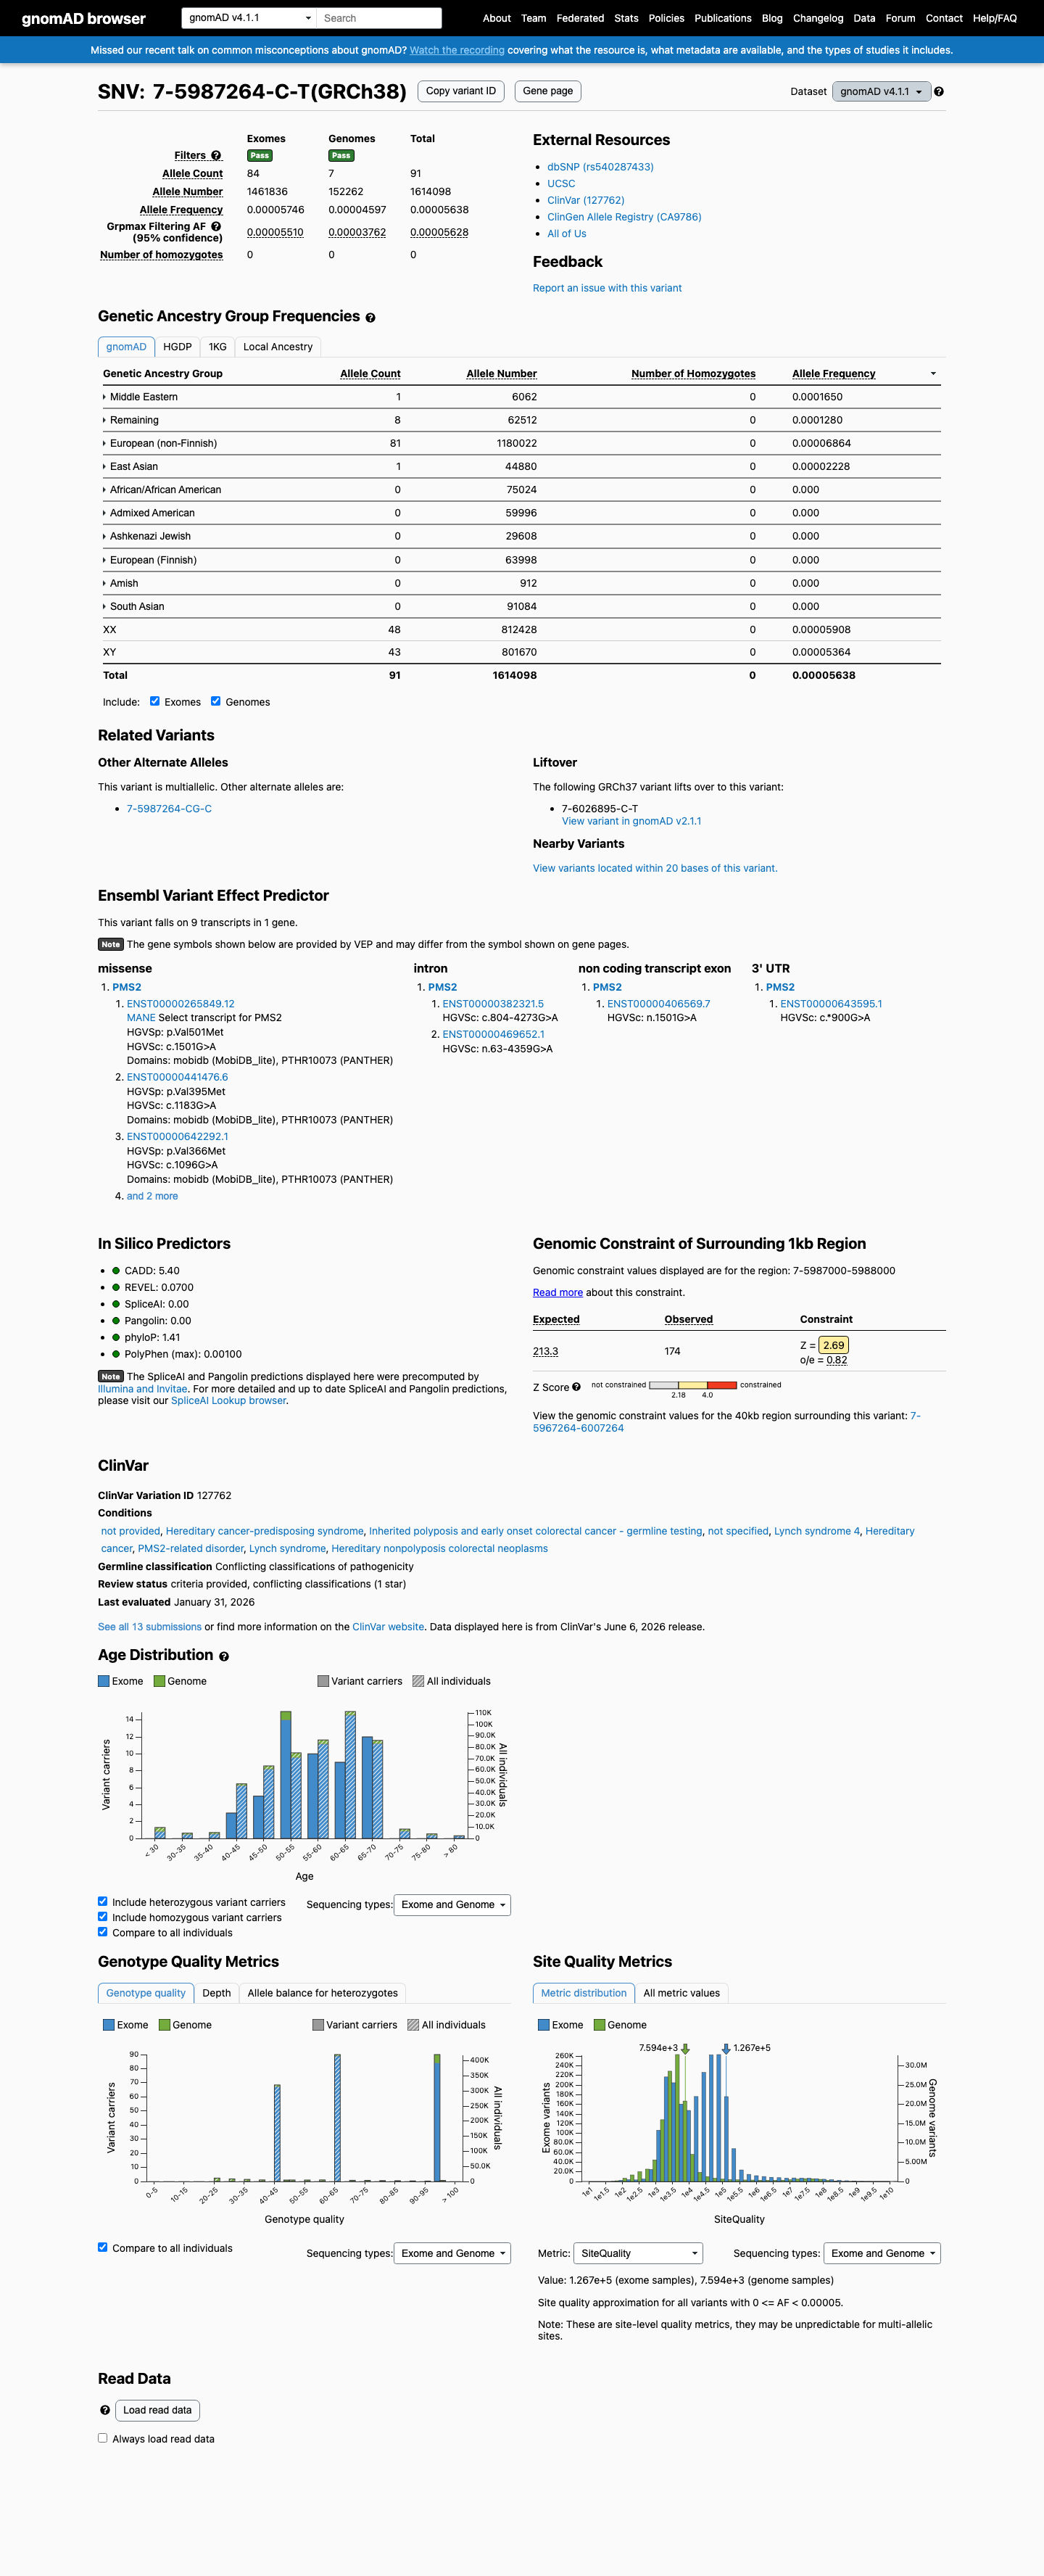
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 5.63782e-05; MAF= 0.00564%, 91/1614098 alleles, homozygotes = 0) and has highest observed frequency in the Middle Eastern population (AF= 0.000164962; MAF= 0.01650%, 1/6062 alleles, homozygotes = 0); grpmax FAF= 5.628e-05.
v2.1
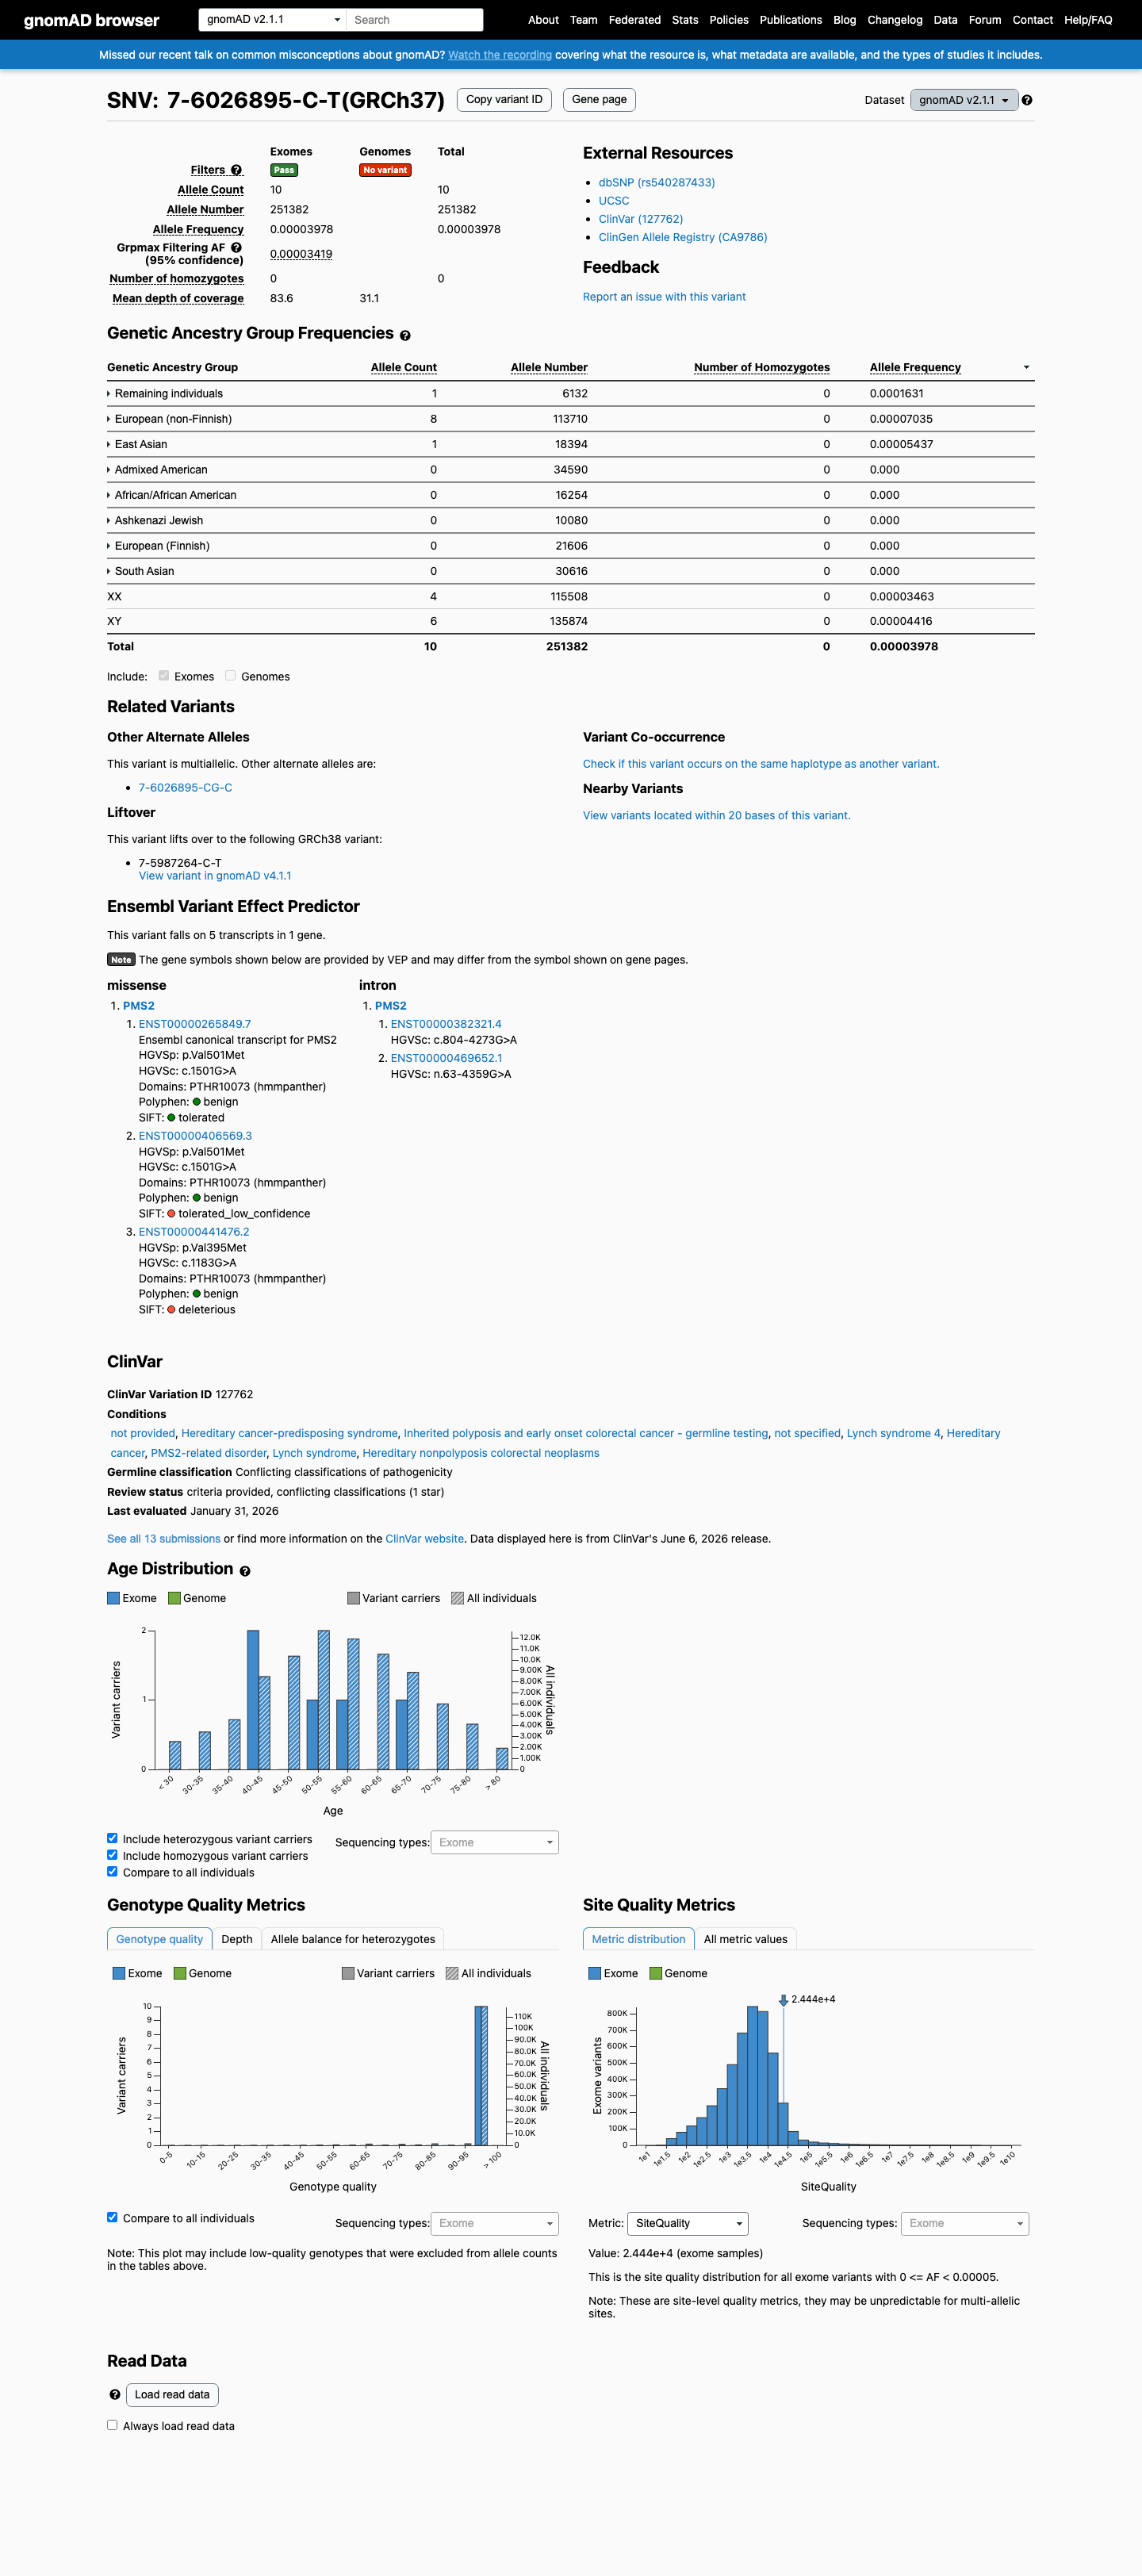
This variant is present in gnomAD v2.1 (AF= 3.97801e-05; MAF= 0.00398%, 10/251382 alleles, homozygotes = 0) and has highest observed frequency in the Remaining individuals population (AF= 0.000163079; MAF= 0.01631%, 1/6132 alleles, homozygotes = 0); grpmax FAF= 3.419e-05.
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.0056% · 91 / 1,614,098
0 hom · FAF 0.0056%
Middle Eastern
1 / 6,062
0.016%
Remaining individuals
8 / 62,512
0.013%
European (non-Finnish)
81 / 1,180,022
0.0069%
East Asian
1 / 44,880
0.0022%
+ 6 not observed (Admixed American, European (Finnish), Amish, South Asian, Ashkenazi Jewish, African/African American)
gnomAD v2.1
0.004% · 10 / 251,382
0 hom · FAF 0.0034%
Remaining individuals
1 / 6,132
0.016%
European (non-Finnish)
8 / 113,710
0.007%
East Asian
1 / 18,394
0.0054%
+ 5 not observed (African/African American, Admixed American, Ashkenazi Jewish, European (Finnish), South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
ClinVar screenshot
ClinVar
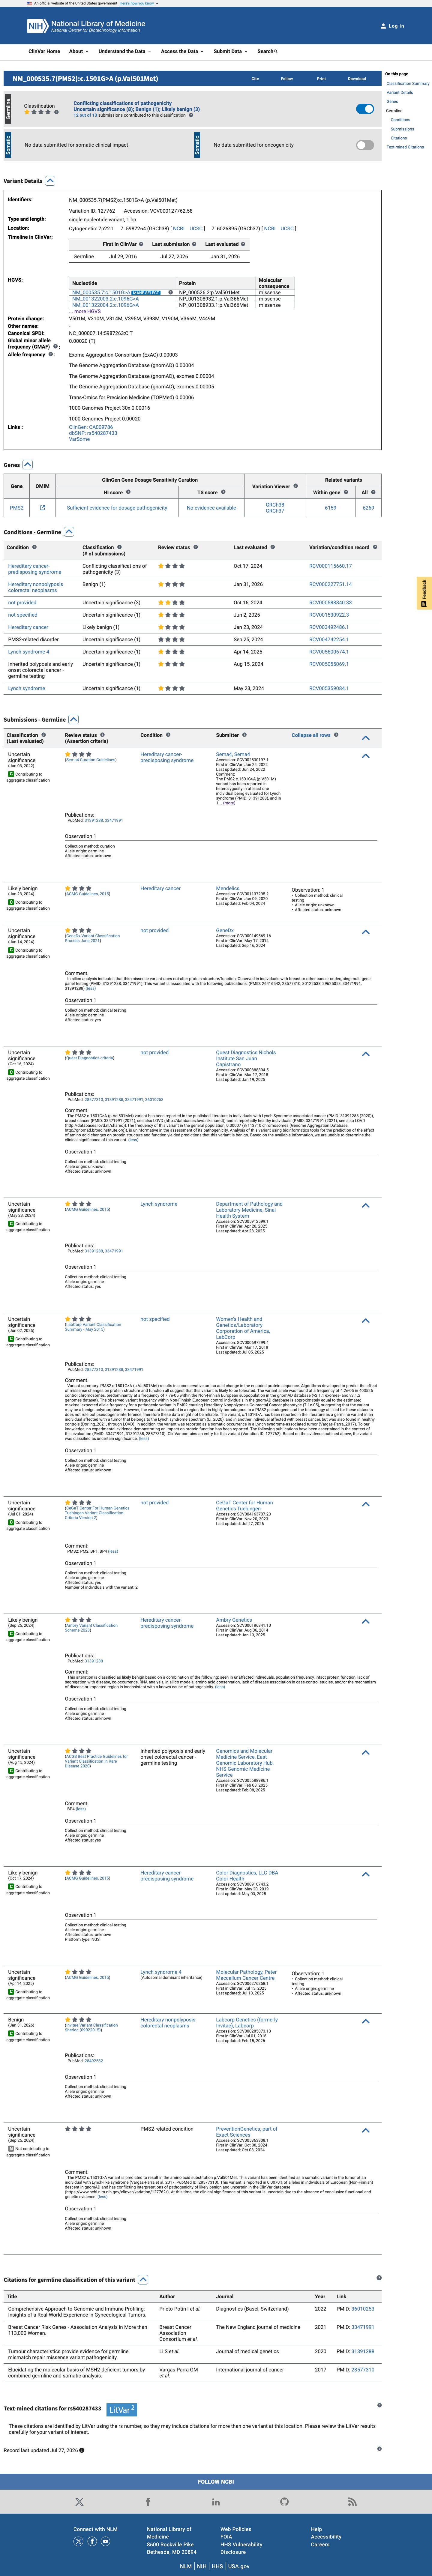
This variant has been reported in ClinVar as Uncertain significance (8 clinical laboratories) and as Likely benign (3 clinical laboratories) and as Benign (1 clinical laboratory). (ClinVarID = 127762)
SpliceAI screenshot
In silico
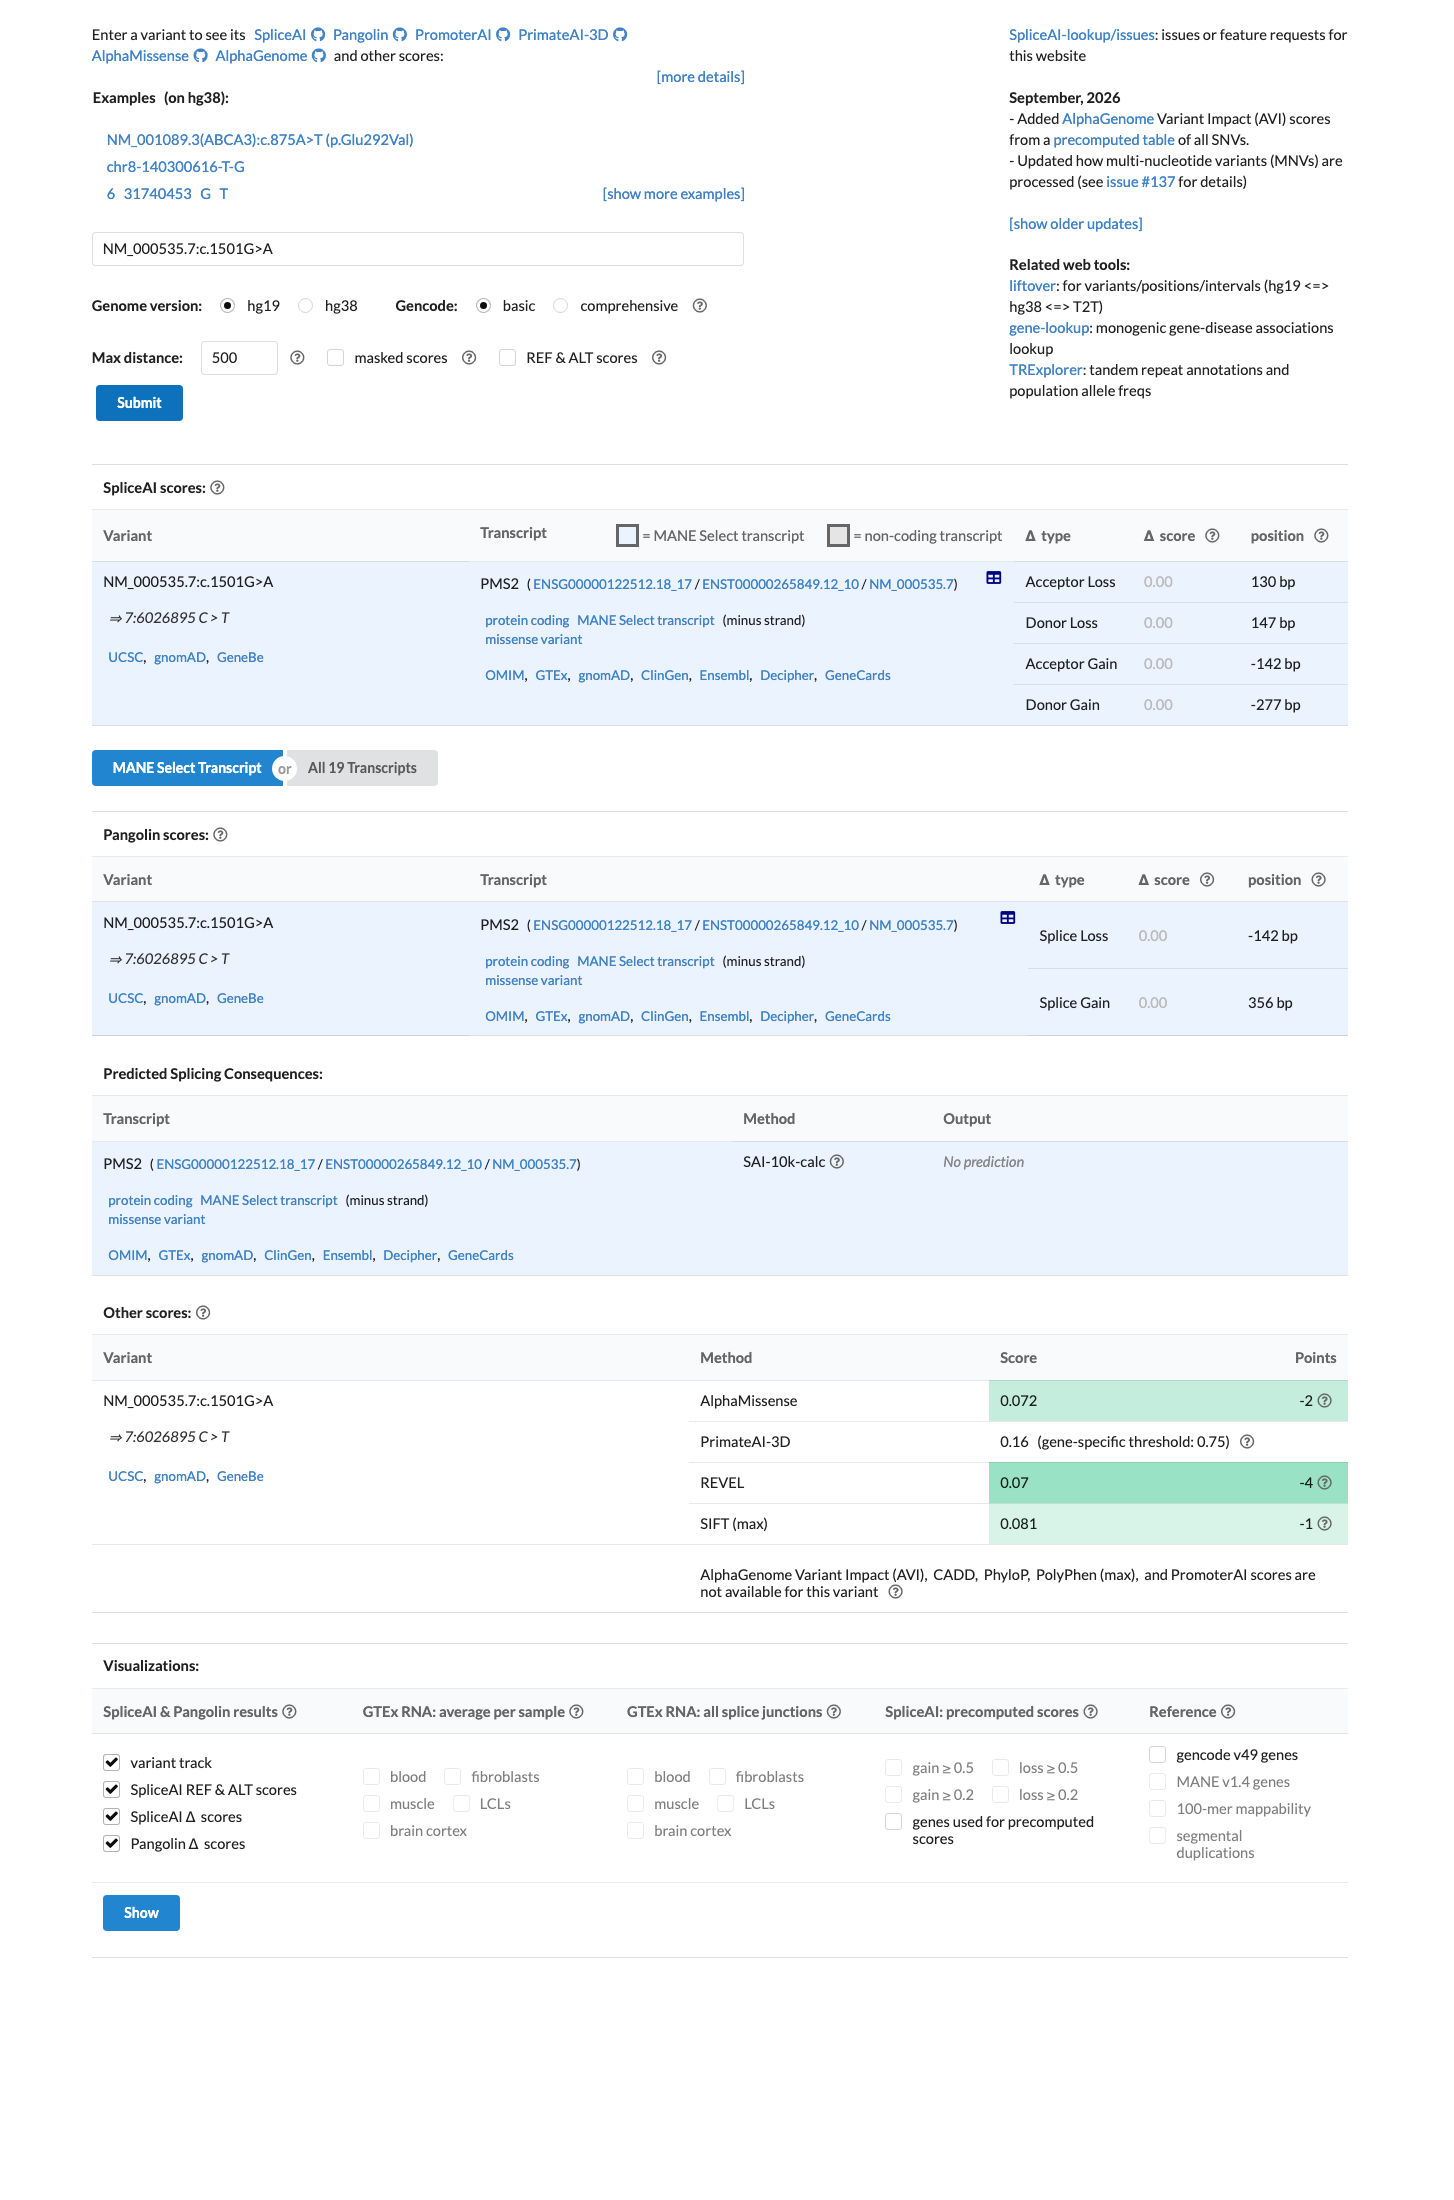
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.00). REVEL score = 0.07. BayesDel score = -0.530405. HCI prior probability for pathogenicity = 0.002. MAPP score = 3.75. Custom PP2 score = 0.001.
Functional / OncoKB screenshot
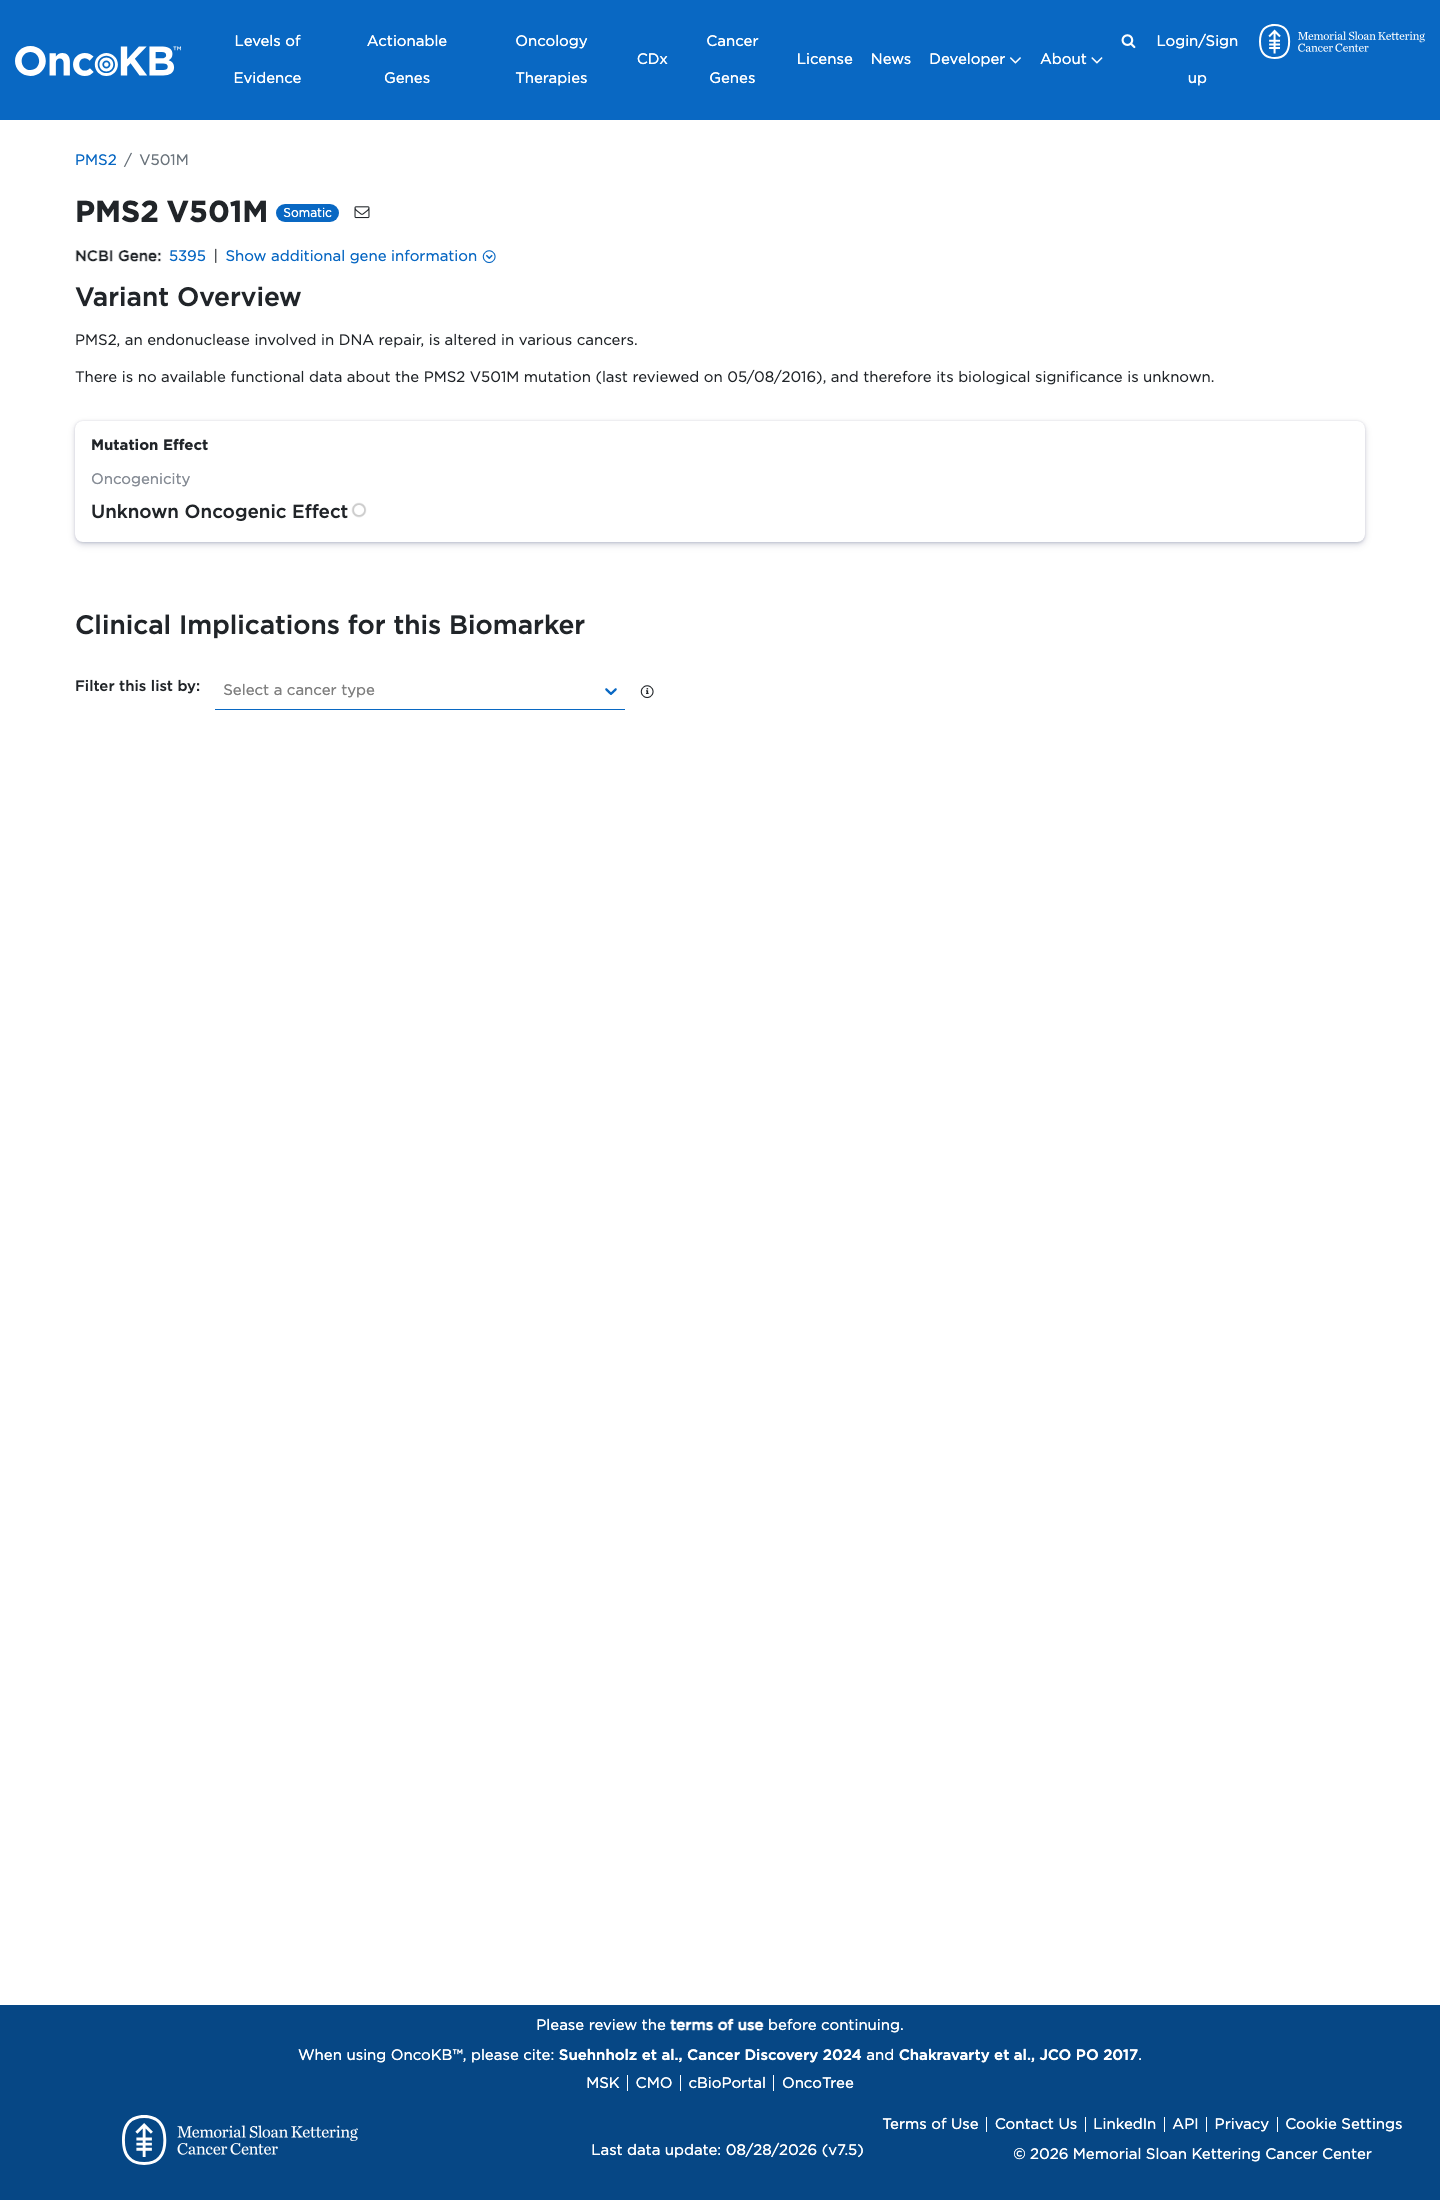
Functional Unknown Oncogenic Effect
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. PMS2, an endonuclease involved in DNA repair, is altered in various cancers.
OncoKB ↗
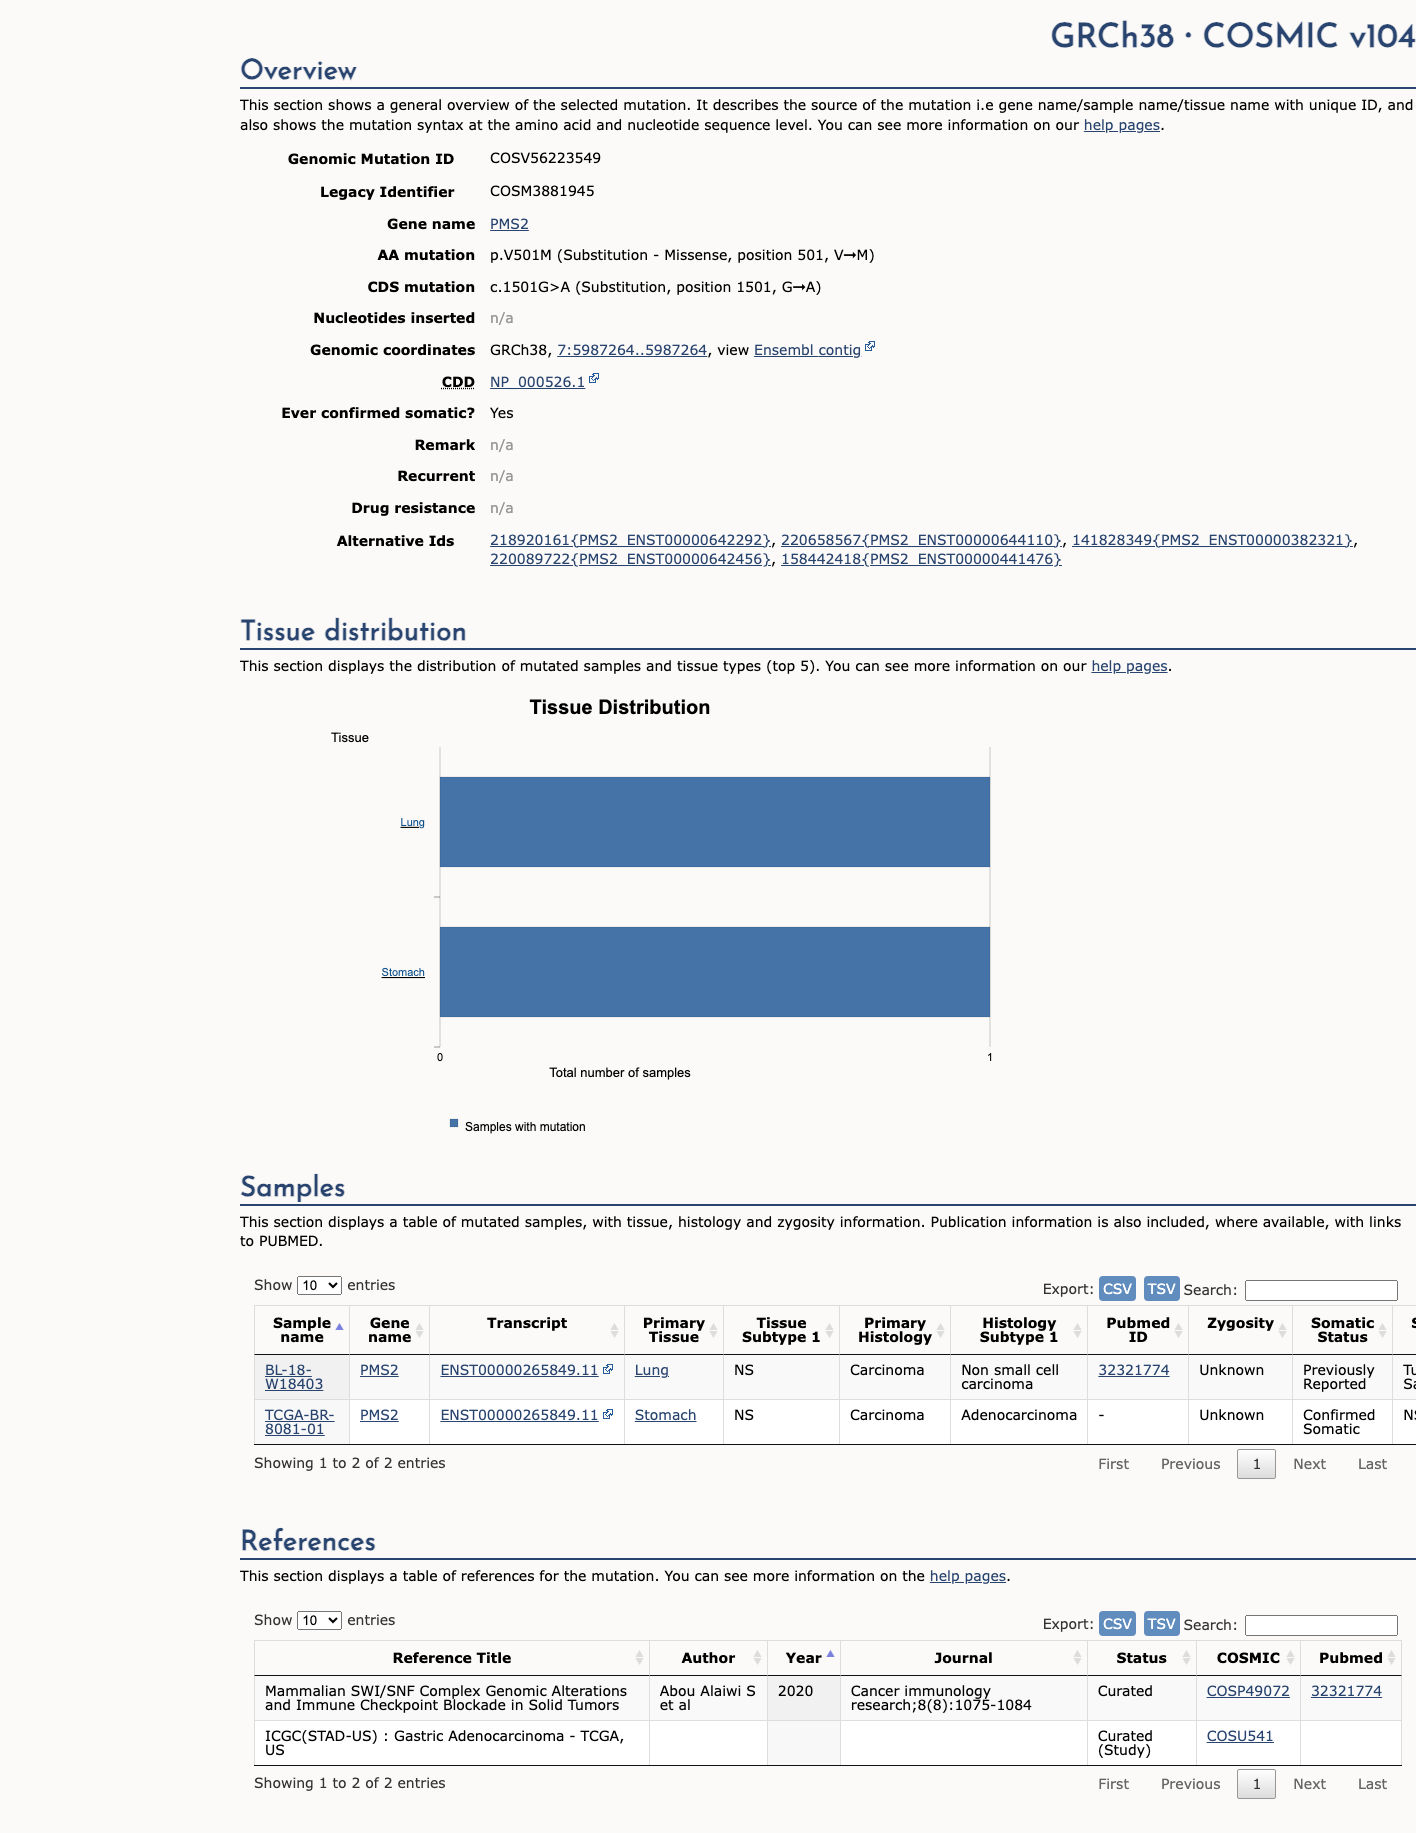
COSMIC screenshot
COSMIC
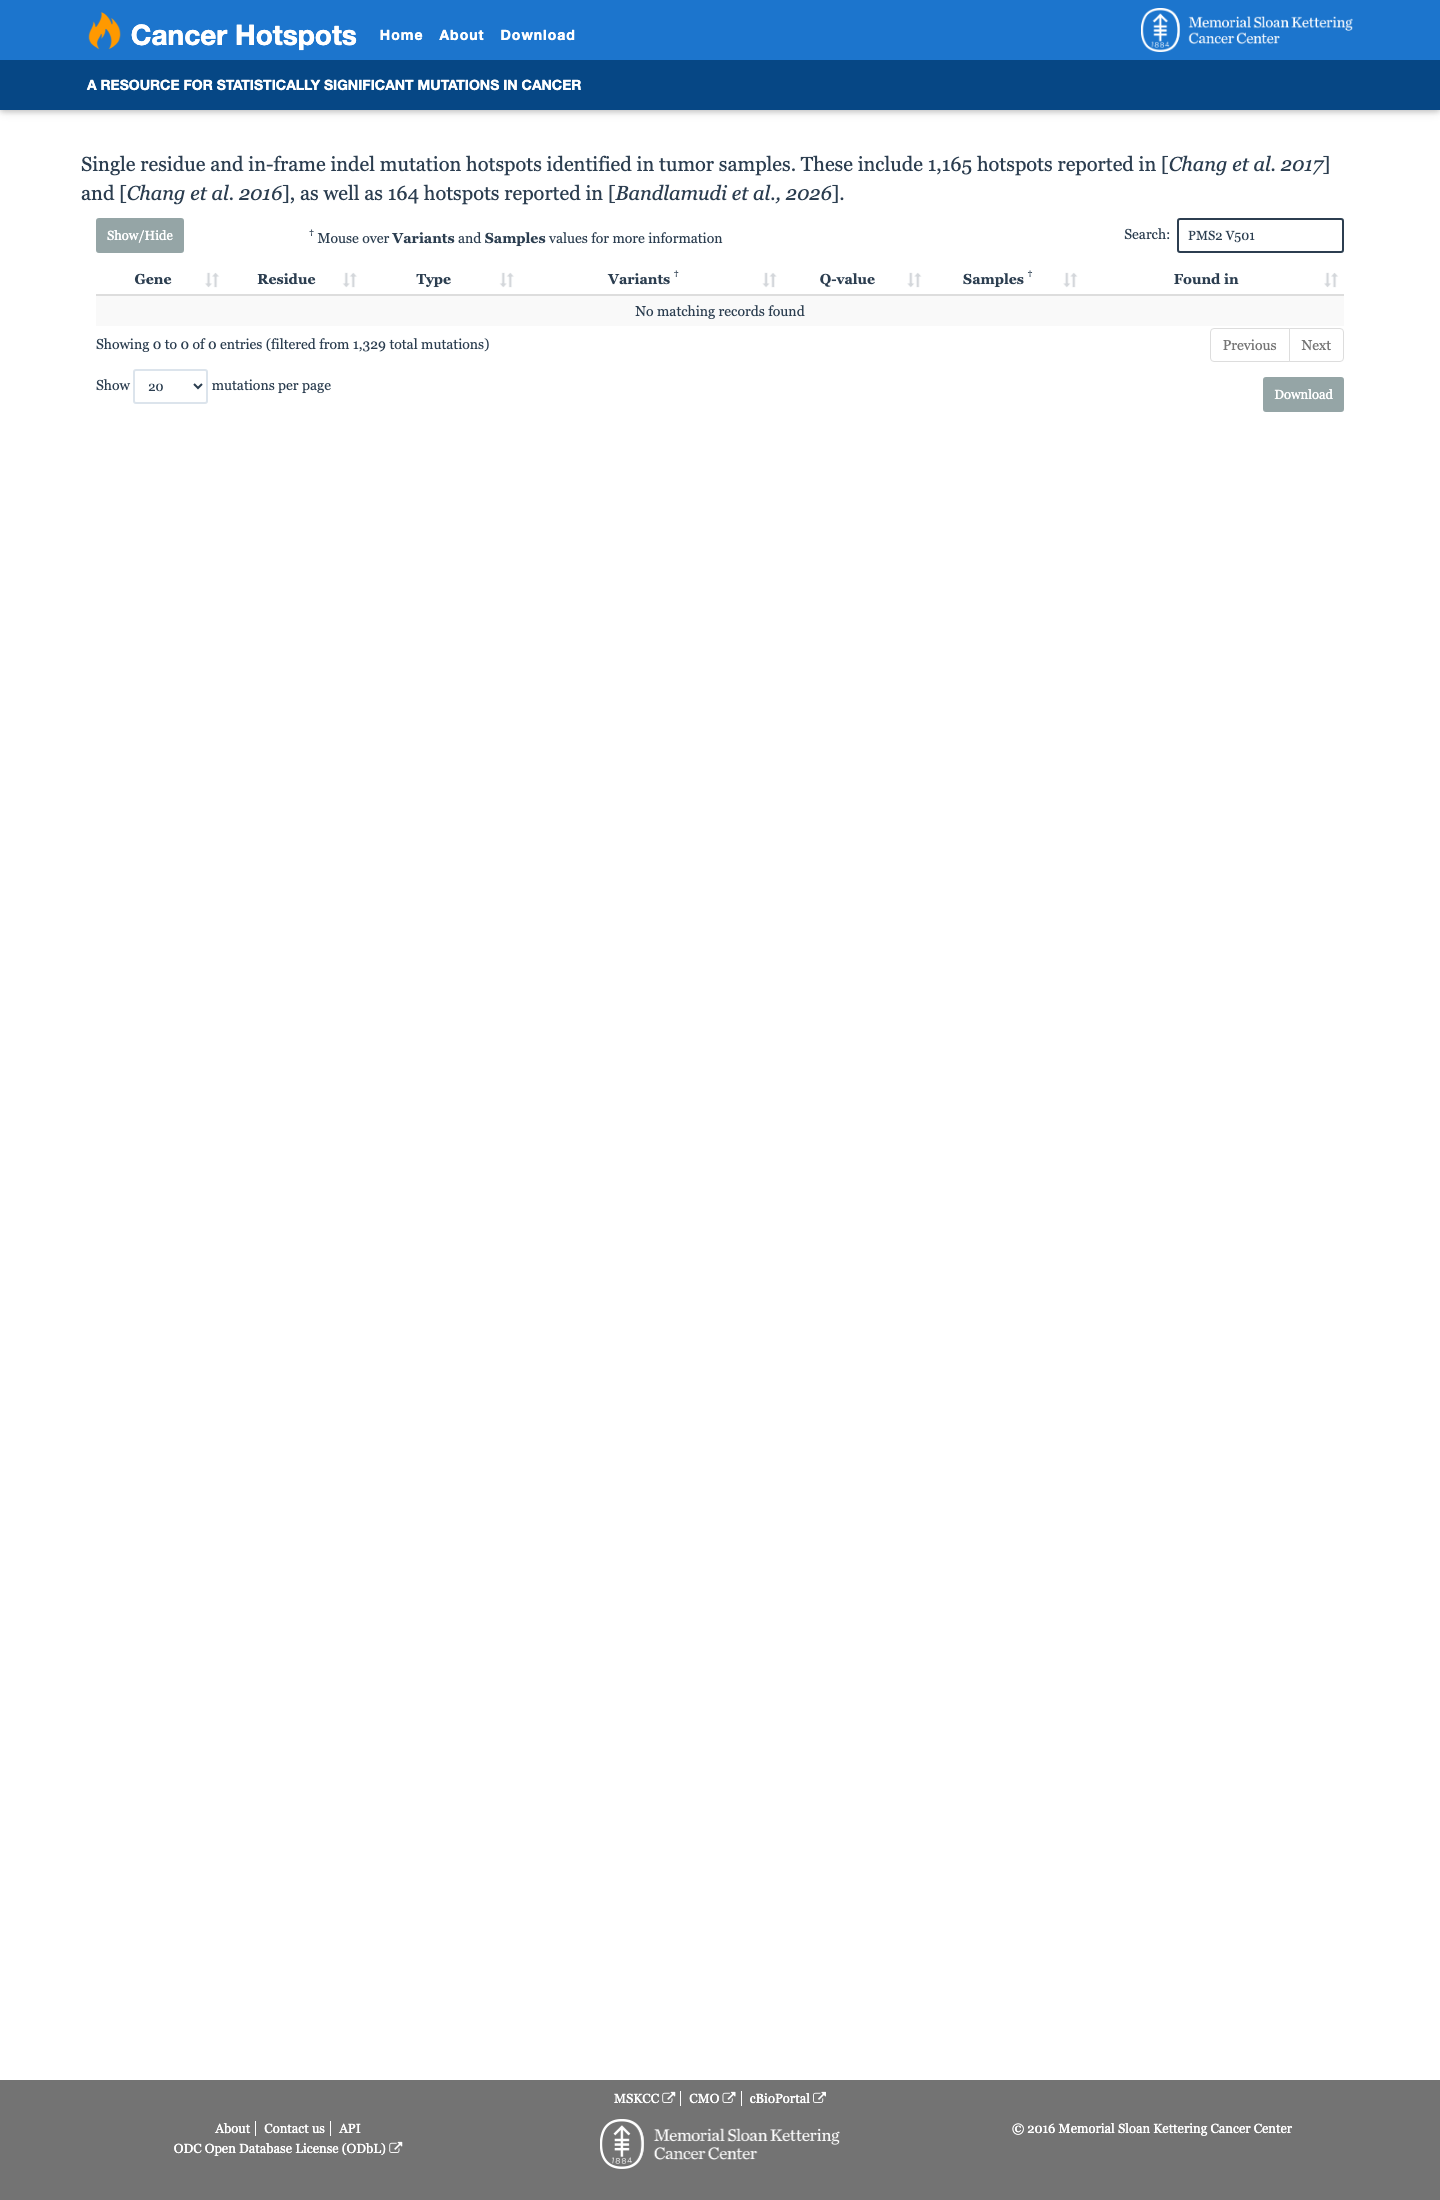
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has previously been reported in somatic cancers (COSMIC; COSV56223549, n = 2 times).
Hotspots
This variant does not lie in a statistically significant hotspot.
Sources & reference links
9Sources
CSpec VCEP
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 8 PMIDs not cited in assessment
15604628 ↗ Genetic cancer risk assessment and counseling: recommendations of the national society of genetic counselors. CLINVAR
20301390 ↗ Lynch Syndrome. CLINVAR
23535968 ↗ Informing family members of individuals with Lynch syndrome: a guideline for cli CLINVAR
25356965 ↗ ACMG policy statement: updated recommendations regarding analysis and reporting CLINVAR
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint co CLINVAR
26467025 ↗ A Standardized DNA Variant Scoring System for Pathogenicity Assessments in Mende CLINVAR
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification crite CLINVAR
28577310 ↗ Elucidating the molecular basis of MSH2-deficient tumors by combined germline and somatic analysis. CLINVAR