Back
NM_000535.7:c.73C>T
p.Gln25Ter · PMS2
0%
complete
Final classification
VUS
PVS1PM2
PMS2
c.73C>T
p.Gln25Ter
nonsense · exon 2

PMS2 is a DNA mismatch-repair protein that works with MLH1 to correct replication errors, including mismatches and small insertion or deletion loops. It acts as a tumor suppressor, and inherited loss of PMS2 function can cause Lynch syndrome and constitutional mismatch repair deficiency, increasing the risk of colorectal, endometrial, ovarian, urothelial, and other cancers. Loss of PMS2 function in tumors can cause hypermutation and microsatellite instability, which may make some cancers responsive to certain immunotherapies.

This variant

This PMS2 variant affects a DNA mismatch-repair tumor-suppressor protein whose inherited loss of function can cause Lynch syndrome and constitutional mismatch repair deficiency.

Transcript
NM_000535.7
HGVS · transcript:coding
NM_000535.7:c.73C>T
GRCh38
chr7:6005982 G>A
GRCh37
chr7:6045613 G>A
VUS: PVS1 (very strong) plus PM2 (supporting) do not satisfy any PMS2 VCEP pathogenic or benign combination rule.
Classification rationale
PVS1PM2 VUS
PMS2 c.73C>T nonsense · exon 2

VUS: PVS1 very strong applies because the nonsense variant truncates PMS2 at codon 25, within the VCEP threshold of codon 798 or earlier. VUS: PM2 supporting applies because the gnomAD v4.1 allele frequency is 6.20819e-07, below 0.00002.

PVS1 + PM2 → VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_000535.7 · variants mapped to exon structure
PMS2 NM_000535.7
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 11 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
PVS1 very strong Pathogenic
Met, very strong: the PMS2 nonsense variant creates a premature stop at codon 25, within the VCEP threshold of codon 798 or earlier.
Case variant facts: NM_000535.7:c.73C>T, NP_000526.2:p.(Gln25Ter)/p.(Q25*), nonsense consequence, PMS2 exon 2.The VCEP PVS1 rule states: Nonsense/frameshift variant introducing Premature Termination Codon (PTC) ≤ codon 798 in PMS2. Refer to Appendix for details.VCEP-pilot-variants---MMR.xlsx.txt was searched for c.73C>T, p.Gln25Ter, and p.Q25*; the matching variant was not listed.
PM2 supporting Pathogenic
Met at Supporting: gnomAD v4 total AF 6.20819e-07 is below 0.00002, with 1 allele in 1,610,776 and no homozygotes.
InSiGHT PMS2 VCEP CSPEC PM2 rule: absent/extremely rare allele frequency <0.00002 (<1 in 50,000 alleles) in the gnomAD v4 dataset.gnomAD v4.1 record for NM_000535.7:c.73C>T (p.Gln25Ter/p.Q25*): total AF 6.208187854797936e-07, 1/1610776 alleles, homozygotes 0; exome AC 1 and exome AN 1458496.VCEP-pilot-variants---MMR.xlsx.txt was searched for c.73C>T, p.Gln25Ter, p.Q25*, Q25*, and Q25Ter; no matching row was found.
Assessed · not applied · 0 not met · 11 not assessed
Pathogenic
PS2 Not assessed: no proband-specific de novo finding, parental testing, or VCEP de novo point total is available for this variant.
PS3 Not assessed: no variant-specific functional assay result is available to compare with the VCEP's calibrated assay criteria or functional-assay flowchart.
PM3 Not assessed: no affected-proband observation, pathogenic PMS2 allele, or confirmed trans/cis phase is documented to generate a PM3 point total.
PP1 Not assessed: no variant-specific pedigree, relative genotypes, meioses, or combined Bayes Likelihood Ratio is available for PP1.
PP4 Not assessed: no patient-specific MSI, tumor-genome, or PMS2 IHC result is available to compare with the VCEP PP4 tumor-evidence rule.
Benign
BA1 Not assessed: gnomAD v4 total AF is 6.20819e-07, but the VCEP requires a Grpmax filtering AF, which is unavailable.
BS1 Not assessed: gnomAD v4 total AF is 6.20819e-07, but the VCEP requires a Grpmax filtering AF, which is unavailable.
BS2 Not assessed: no qualifying in-trans co-occurrence, cancer-age, CMMRD-exclusion, or phase-confirmation evidence is available.
BS3 Not assessed: no variant-specific proficient-function result or calibrated functional odds are available for comparison with the VCEP BS3 thresholds.
BS4 Not assessed: no variant-specific non-segregation pedigree, relative data, meioses, or combined Bayes Likelihood Ratio is available for BS4.
BP5 Not assessed: no patient-specific MSS, MMR-protein, BRAF V600E, or MLH1-methylation tumor evidence is available for the BP5 alternate-basis rule.
N/A · 15 PS1 · PS4 · PM1 · PM4 · PM5 · PM6 · PP2 · PP3 · PP5 · BP1 · BP2 · BP3 · BP4 · BP6 · BP7
Research & evidence
Population frequency
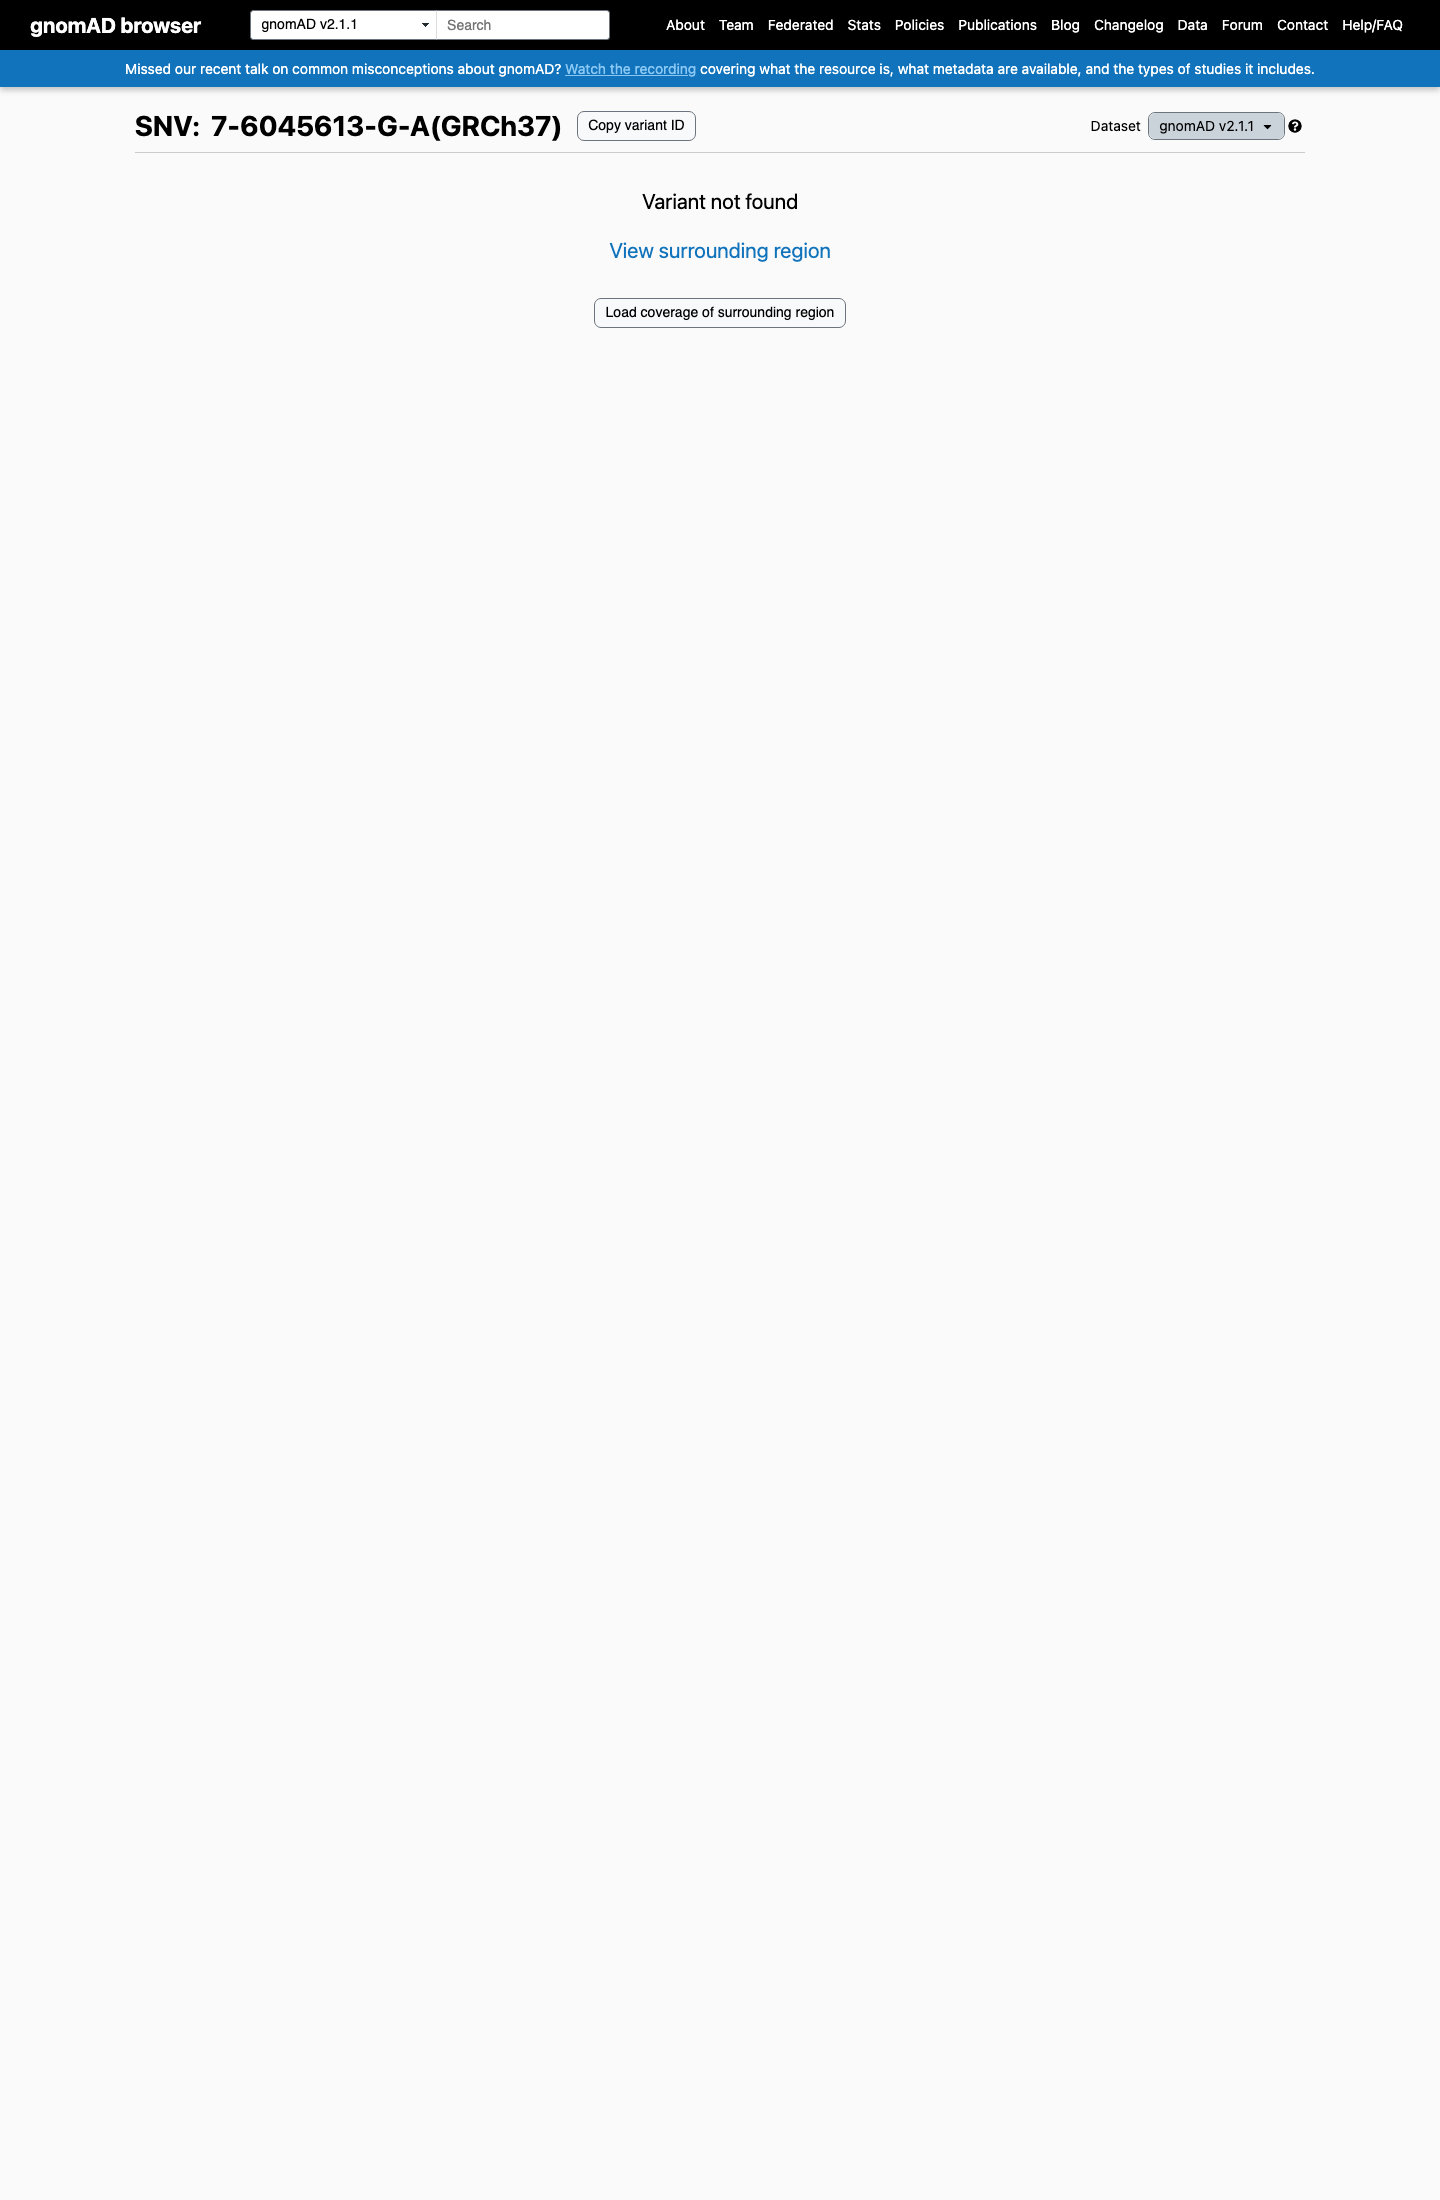
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
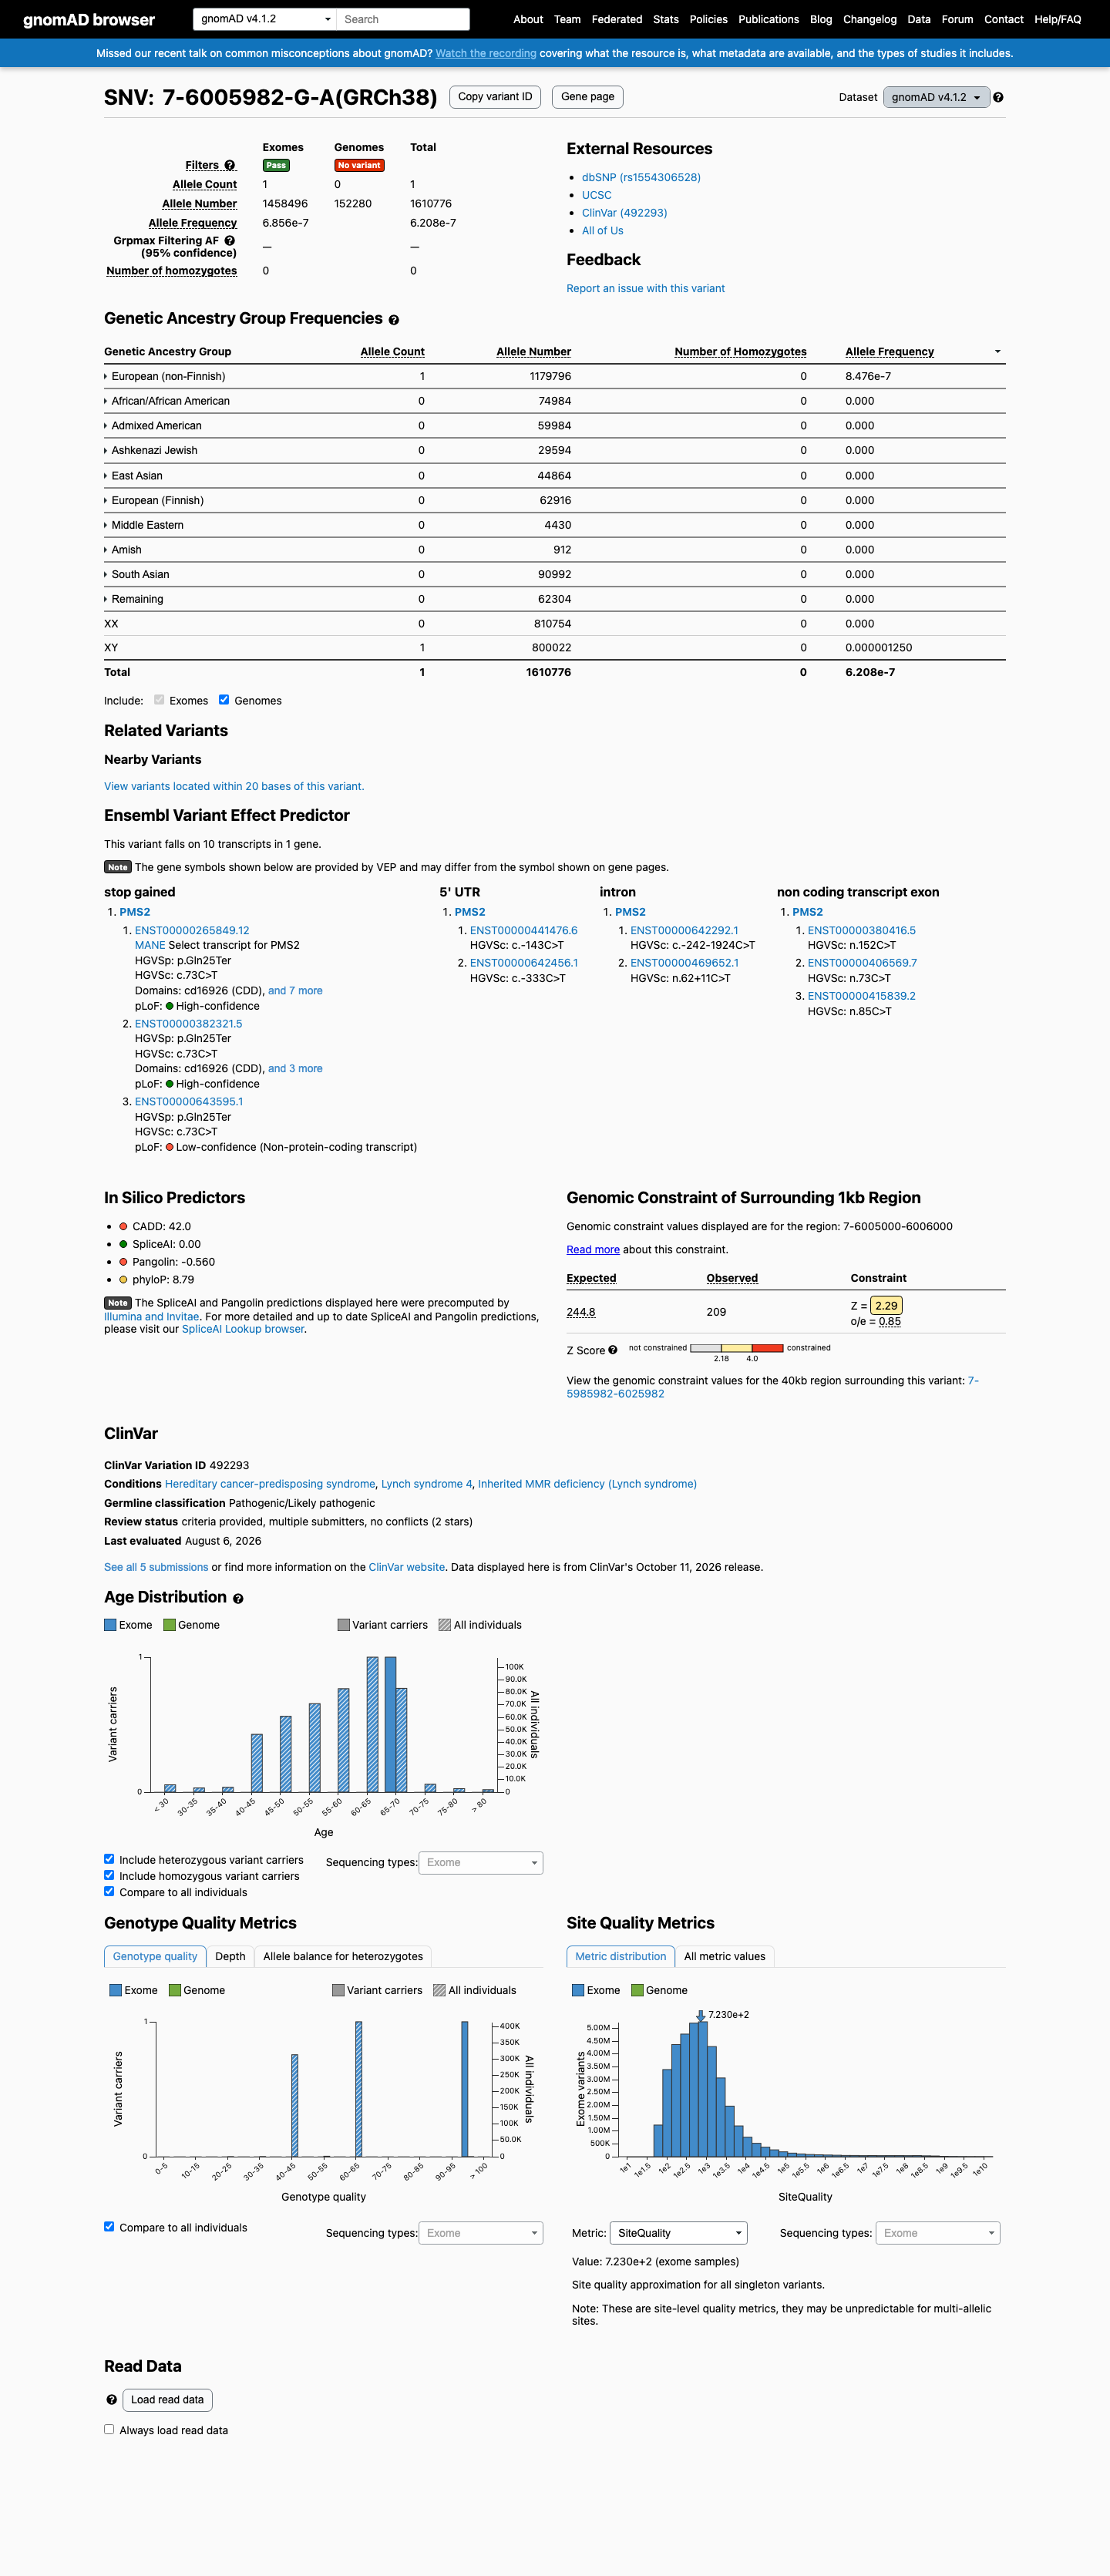
This variant is present in gnomAD v4.1 (AF= 6.20819e-07; MAF= 0.00006%, 1/1610776 alleles, homozygotes = 0) and has highest observed frequency in the European (non-Finnish) population (AF= 8.47604e-07; MAF= 0.00008%, 1/1179796 alleles, homozygotes = 0).
v2.1
Absent from gnomAD v2.1.
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
6.2e-05% · 1 / 1,610,776
0 hom
European (non-Finnish)
1 / 1,179,796
8.5e-05%
+ 9 not observed (Remaining individuals, Admixed American, European (Finnish), Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American)
gnomAD v2.1
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
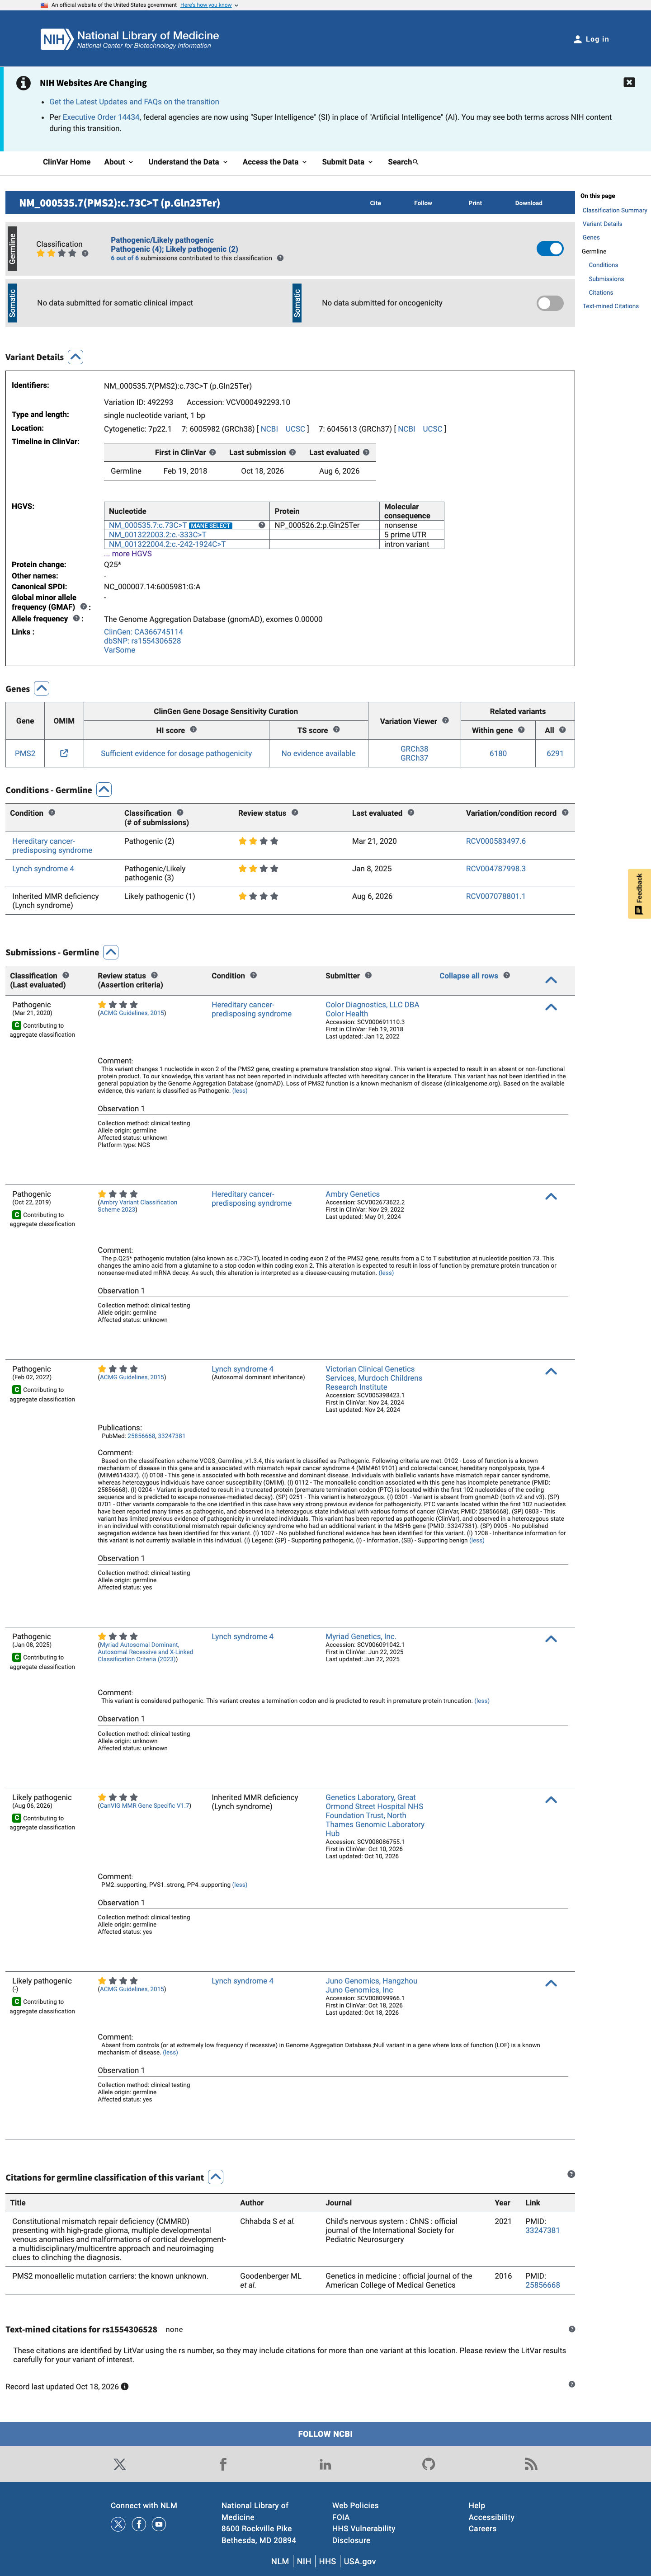
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Pathogenic (4 clinical laboratories) and as Likely pathogenic (2 clinical laboratories). (ClinVarID = 492293)
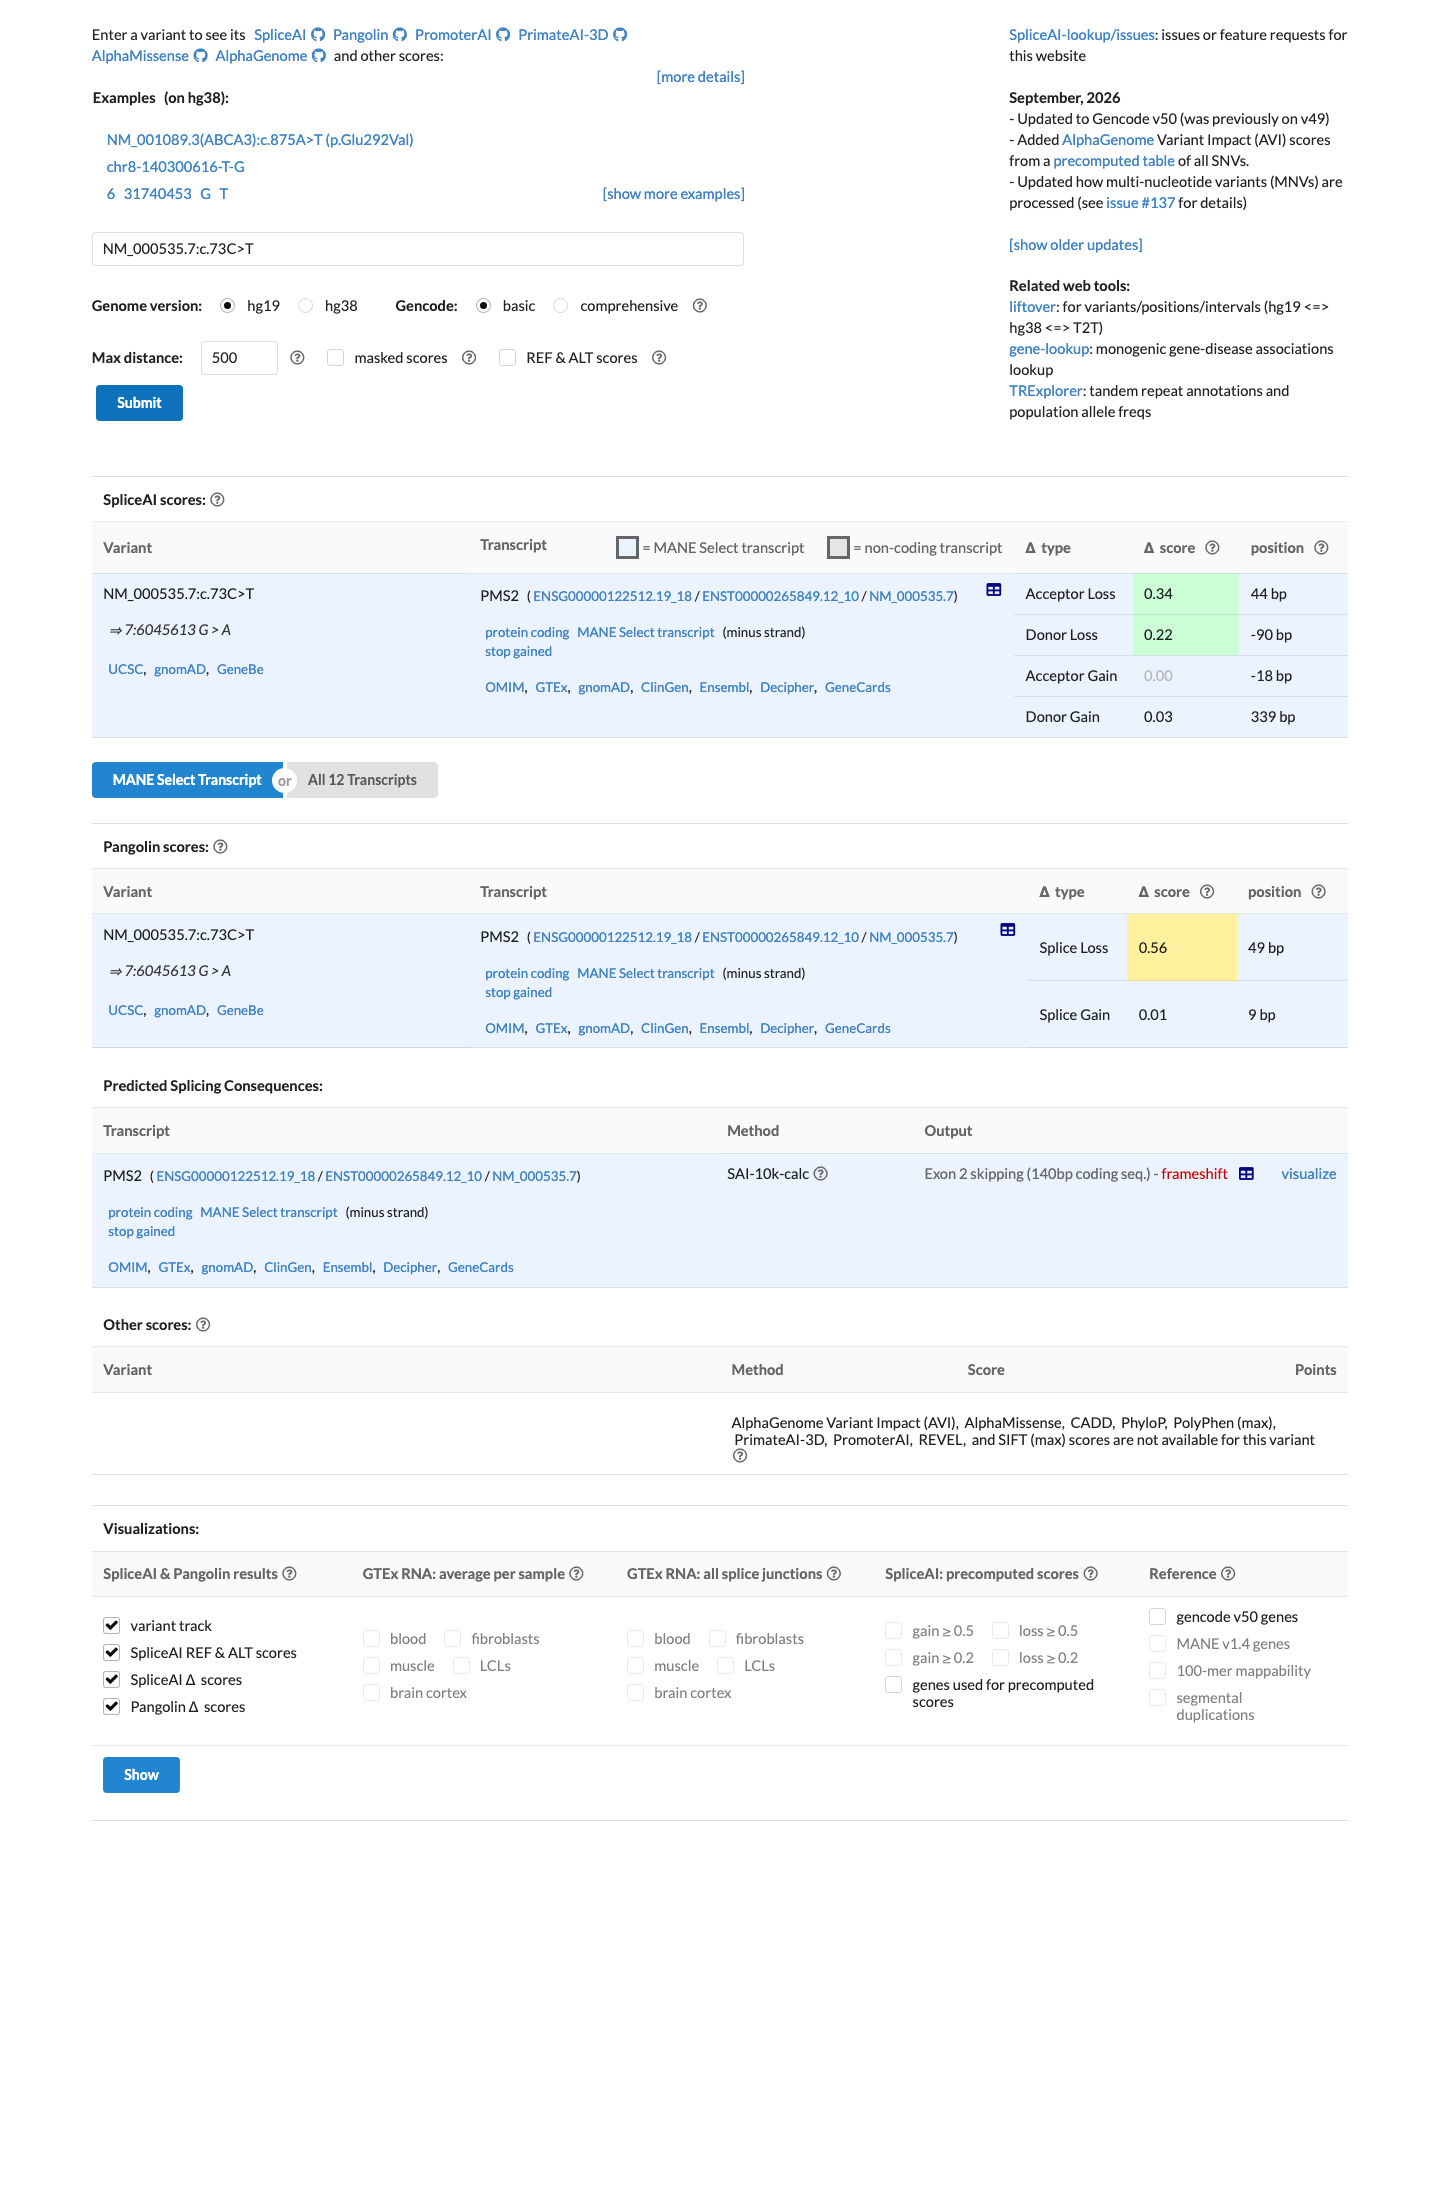
SpliceAI screenshot
In silico
SpliceAI predicts possible splice impact for this variant (max delta score = 0.34). BayesDel score = 0.655873.
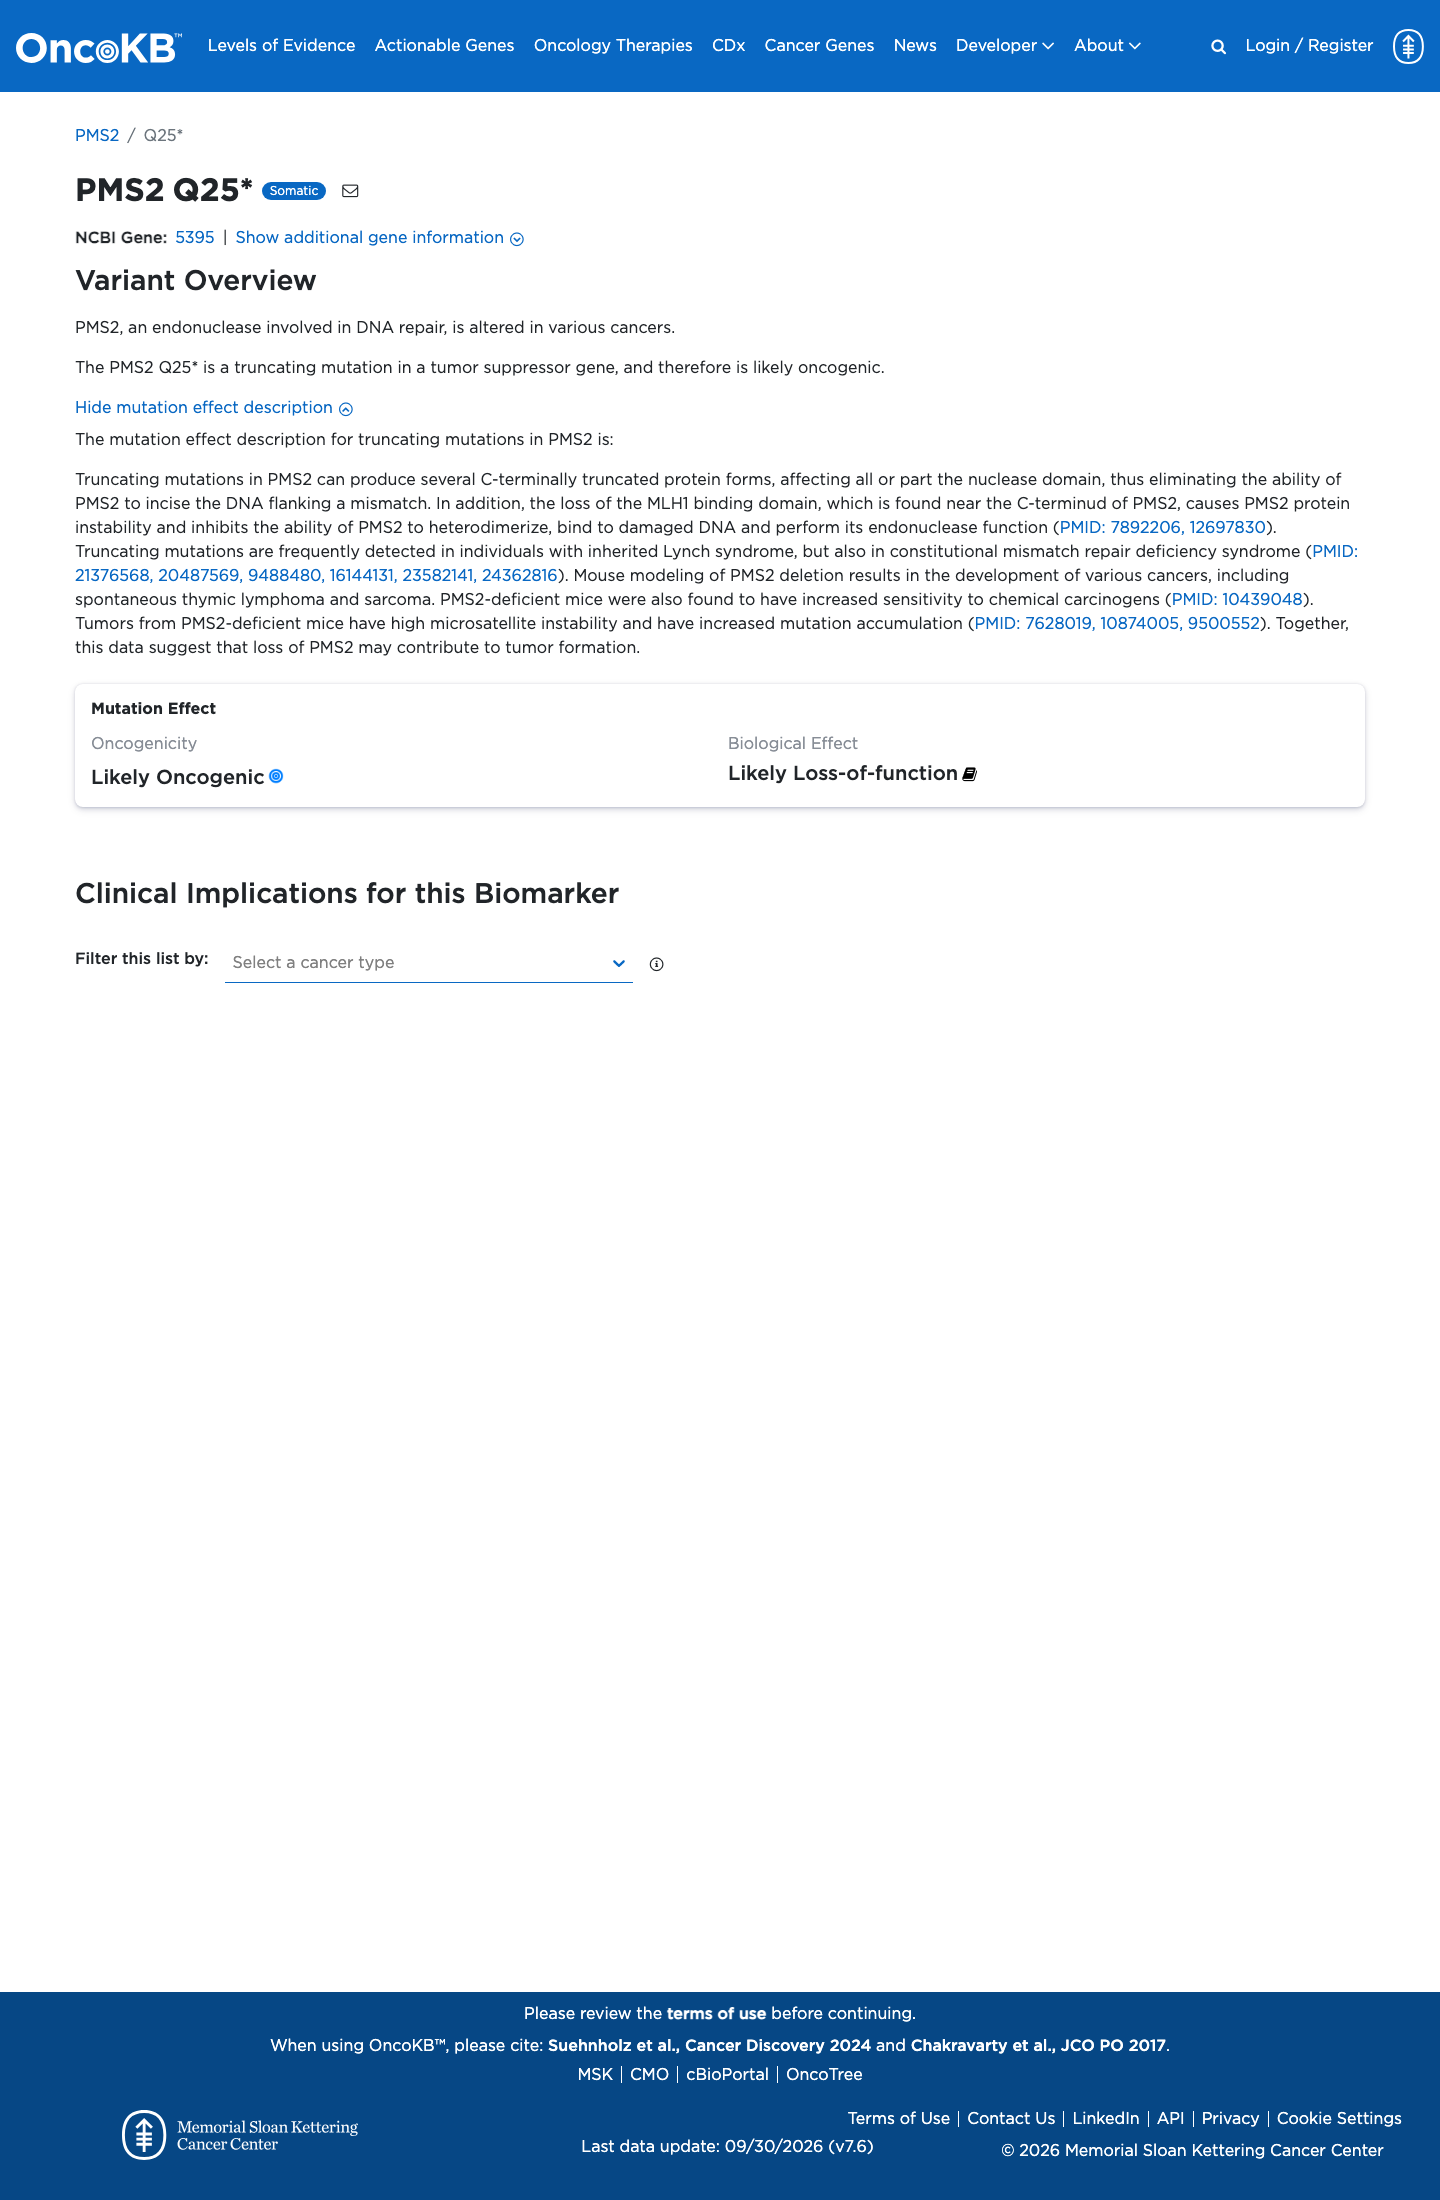
Functional / OncoKB screenshot
Functional Likely Oncogenic
OncoKB identified variant-specific curated literature and context relevant to functional review; biological-effect context: Likely Loss-of-function; curated oncogenicity label: Likely Oncogenic.
OncoKB ↗
COSMIC screenshot
COSMIC
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
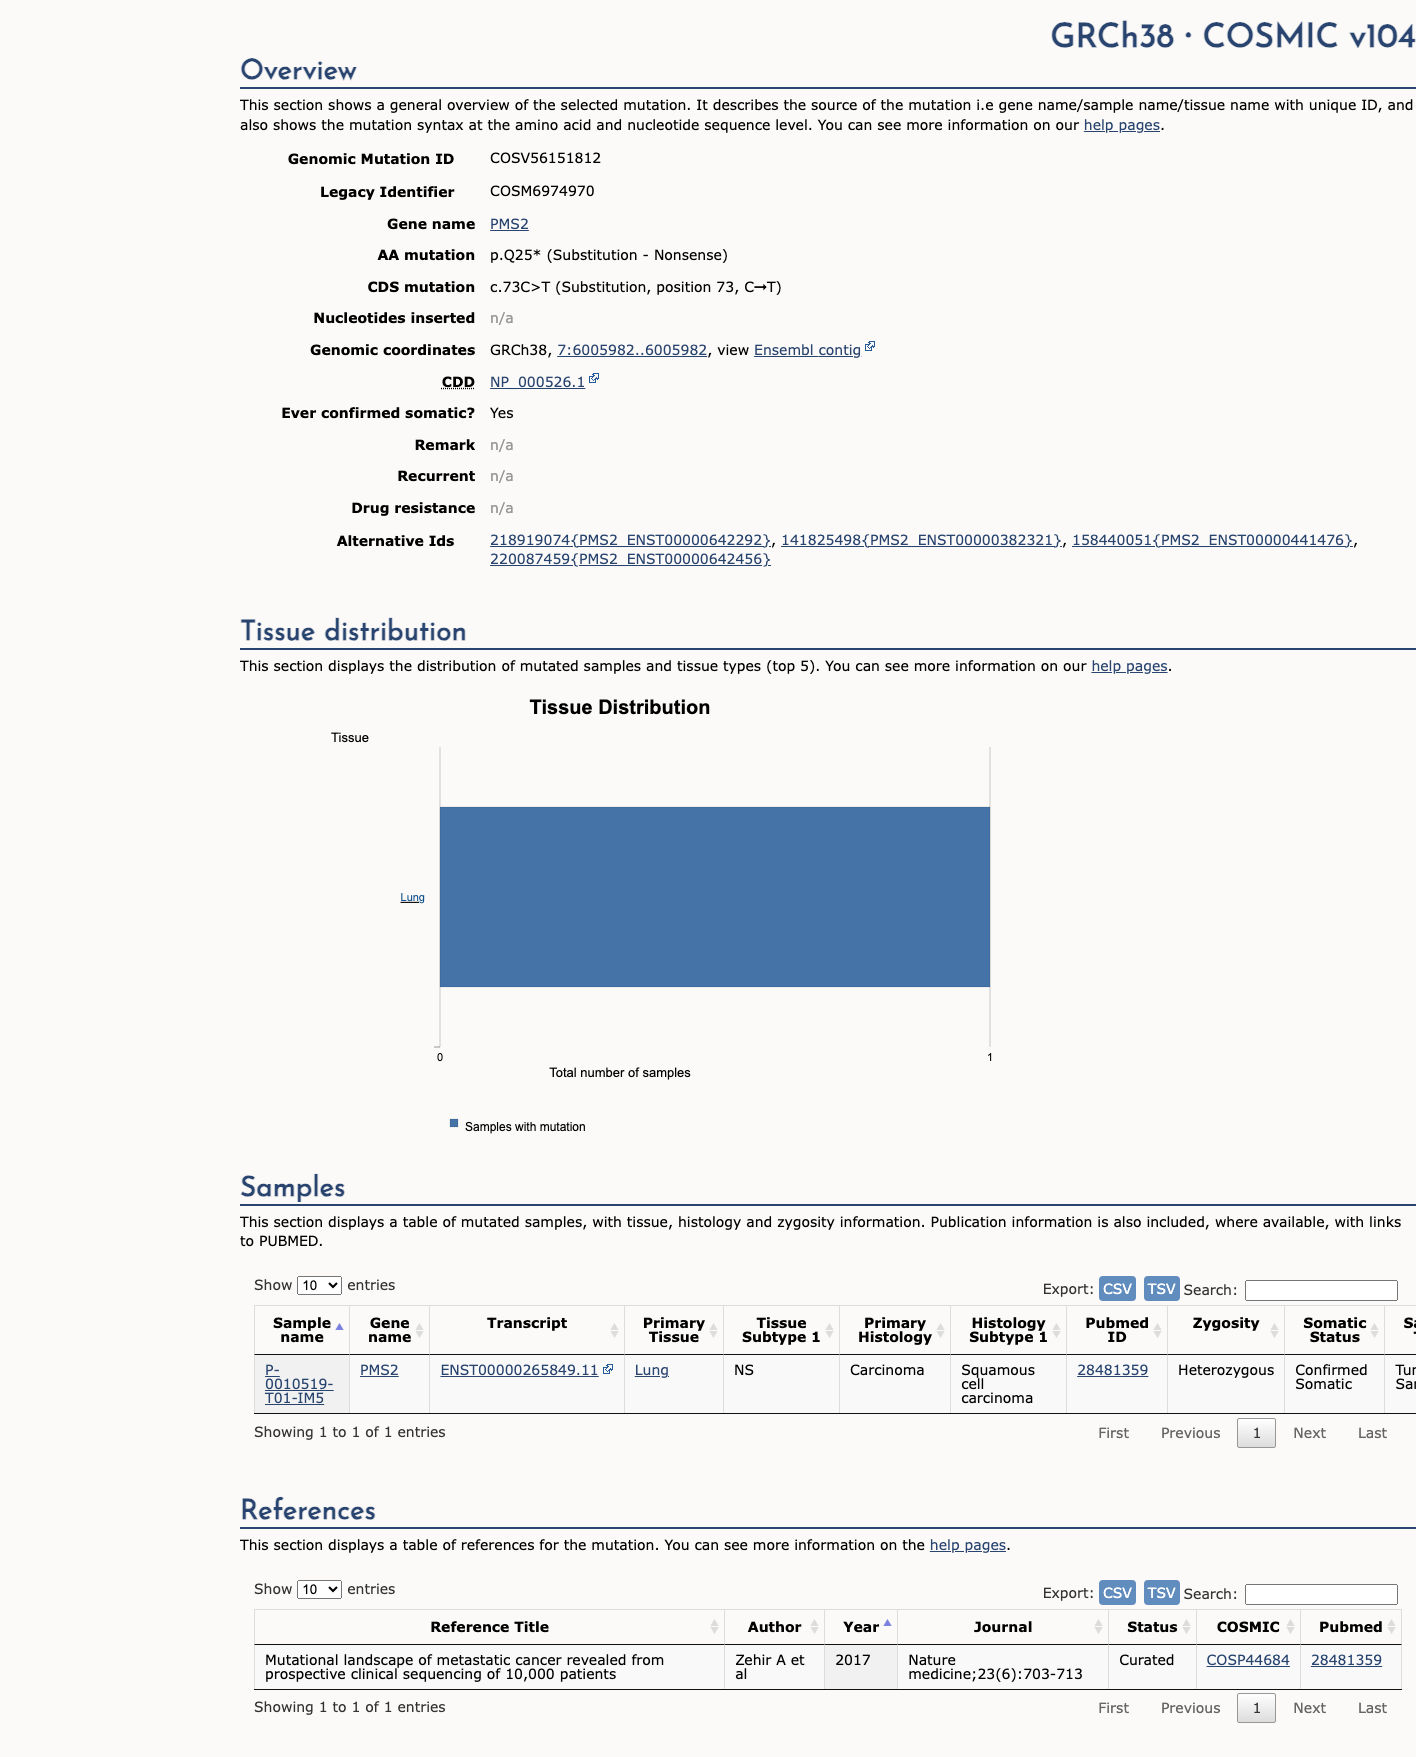
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has previously been reported in somatic cancers (COSMIC; COSV56151812, n = 1 times).
Hotspots
This variant does not lie in a statistically significant hotspot.
Sources & reference links
9Sources
CSpec VCEP
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 8 PMIDs not cited in assessment
10439048 ↗ Mice defective in the DNA mismatch gene PMS2 are hypersensitive to MNU induced thymic lymphoma and are partially protected by transgenic expression of human MGMT. ONCOKB
10874005 ↗ Mutagenesis in PMS2- and MSH2-deficient mice indicates differential protection from transversions and frameshifts. ONCOKB
12697830 ↗ Dimerization of MLH1 and PMS2 limits nuclear localization of MutLalpha. ONCOKB
16144131 ↗ Two PMS2 mutations in a Turcot syndrome family with small bowel cancers. ONCOKB
20487569 ↗ MSH6 and PMS2 mutation positive Australian Lynch syndrome families: novel mutations, cancer risk and age of diagnosis of colorectal cancer. ONCOKB
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology. CLINVAR
20301390 ↗ Lynch Syndrome. CLINVAR
25856668 ↗ PMS2 monoallelic mutation carriers: the known unknown. CLINVAR