Back
NM_001040108.2:c.2390G>A
p.Arg797His · MLH3
ACMG/AMP
0%
complete
Final classification
Benign
BA1BS1BP1BP4
MLH3
c.2390G>A
p.Arg797His
missense · exon 2

MLH3 is a member of the MutL-homolog family of DNA mismatch repair genes. It partners with other family members, notably MLH1, to form a complex that detects and repairs errors in DNA during replication, safeguarding genomic stability and preventing microsatellite instability, and it also helps promote meiotic crossover. Germline mutations in MLH3 are associated with hereditary nonpolyposis colorectal cancer type 7 (HNPCC7), and inactivating changes in the gene have also been found in endometrial and gastric cancers. Loss of MLH3 function increases susceptibility to tumors such as gastrointestinal cancers, indicating that it acts as a tumor suppressor.

This variant

MLH3 is a MutL-homolog mismatch-repair gene whose loss of function, mainly through truncating alleles in a biallelic or HNPCC7-associated setting, predisposes to colorectal and other gastrointestinal cancers; this common missense substitution is tolerated in the general population and is not the type of change expected to abrogate MLH3 tumour-suppressor function.

Transcript
NM_001040108.2
HGVS · transcript:coding
NM_001040108.2:c.2390G>A
GRCh38
chr14:75047266 C>T
GRCh37
chr14:75513969 C>T
Benign: BA1 (stand-alone benign, popmax filtering AF 0.063-0.065) alone satisfies the generic ACMG/AMP 2015 rule, with BS1 (strong) concordant.
Classification rationale
BA1BS1BP1BP4 Benign
MLH3 c.2390G>A missense · exon 2

BA1 stand-alone benign: the allele reaches a popmax filtering frequency of 0.0633 (gnomAD v2.1) and 0.0647 (gnomAD v4.1) in African ancestry, above 5%. BS1 strong: that same 0.063-0.065 popmax filtering frequency is well above the 0.01 strong benign threshold. BP1 supporting: MLH3 disease alleles are predominantly truncating, with 2,175 of 2,176 ClinVar missense variants uncertain, conflicting or benign. BP4 moderate: REVEL 0.083 places this missense substitution at or below the moderate benign in-silico threshold.

BA1 + BS1 + BP1 + BP4 Benign
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_001040108.2 · variants mapped to exon structure
MLH3 NM_001040108.2
Fetching transcript structure from UCSC…
Applied criteria · 4 applied · 19 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 4
Strength Supporting Moderate Strong Very strong
BA1 stand-alone Benign
Met, stand-alone: the African-ancestry popmax filtering AF is 0.0633 in gnomAD v2.1 and 0.0647 in v4.1, both above the 0.05 BA1 threshold.
Governing framework: no CSPEC/VCEP or local MLH3 framework exists (case_summary.json cspec.found = false; review_manifest.json gene_vcep_dir null; vcep_materials.json no CSPEC URL and no gene VCEP files; final_classification_framework.json source = generic_acmg_fallback), so the generic ACMG/AMP 2015 framework governs (PMID:25741868).BA1 threshold applied verbatim from the supplied generic calibration: allele frequency >= 0.05 = stand-alone benign support (ACMG/AMP 2015 stand-alone allele-frequency criterion, PMID:25741868).gnomAD v2.1 (gnomad_v2): total AF 0.006369 (1801/282780 alleles, 72 homozygotes); African/African American AF 0.066113 (1649/24942, 71 homozygotes); grpmax filtering AF 0.063292; exome-only AF 0.004881 (1227/251392, 49 homozygotes) with exome grpmax filtering AF 0.063292; genome-only AF 0.018287 (574/31388, 23 homozygotes).
BS1 strong Benign
Met at strong: the popmax filtering AF of 0.063-0.065 exceeds the 0.01 BS1 threshold, on the same observation that already drives the stand-alone BA1 call.
Governing framework: generic ACMG/AMP 2015 (no CSPEC/VCEP or local MLH3 framework), under which BS1 is strong benign support when the control-population frequency is greater than expected for the disorder (Richards et al. 2015, PMID:25741868).BS1 threshold applied verbatim from the supplied generic calibration: AF >= 0.01 (generic default with per-gene derivation preferred; Whiffin et al. 2017, PMID:28518168).gnomAD v2.1 (gnomad_v2): African/African American AF 0.066113 (1649/24942) with grpmax filtering AF 0.063292; total AF 0.006369 (1801/282780, 72 homozygotes); genome-only AF 0.018287; exome-only AF 0.004881 with exome grpmax filtering AF 0.063292.
BP1 supporting Benign
Met at supporting: MLH3 disease alleles are predominantly truncating (nonsense, frameshift, splice), with 2,175 of 2,176 ClinVar missense variants uncertain, conflicting or benign.
ClinVar MLH3 pathogenic/likely pathogenic records sampled for this assessment are all protein-truncating or splice-disrupting: c.22G>T (p.Glu8Ter), c.239C>A (p.Ser80Ter), c.980dup (p.Ile328fs), c.1852del (p.Thr618fs), c.2903del (p.Thr968fs), c.3112_3115del (p.Asn1038fs), c.3571-1G>A, c.3759del (p.Lys1253fs), c.3958A>T (p.Arg1320Ter), c.3828G>A (p.Trp1276Ter).Missense spectrum: of 2,176 MLH3 missense-annotated ClinVar records, 2,175 are uncertain significance, conflicting, likely benign, benign/likely benign or unclassified; the single P/LP missense-class allele is the initiator-codon change NM_001040108.2:c.2T>C (p.Met1Thr), which is not a classical missense mechanism.Gene-level mechanism (pvs1_gene_context): a biallelic germline nonsense variant of MLH3 underlies polyposis predisposition, and MLH3 is curated as a tumour suppressor whose inactivating changes are associated with HNPCC7, endometrial and gastric cancers - i.e. loss of function, not missense substitution, is the documented disease mechanism.
BP4 moderate Benign
Met at moderate strength: REVEL 0.083 sits at or below the <=0.183 moderate BP4 threshold for this missense variant.
Variant consequence: missense, NP_001035197.1:p.(Arg797His) from NM_001040108.2:c.2390G>A, establishing that the REVEL protein-impact path governs BP4 (the SpliceAI path applies only to intronic/synonymous/splice-region variants).Local REVEL v1.3 lookup: revel_score = 0.083 at chr14:75047266 C>T (hg38; = 14-75513969-C-T hg37); 0.083 <= 0.183 (moderate) and <= 0.290 (supporting) but > 0.016, so BP4 reaches moderate and not strong.Thresholds applied verbatim from the supplied ClinGen SVI REVEL calibration (Pejaver et al. 2022, Am J Hum Genet, PMID:36413997): BP4 supporting <=0.290, moderate <=0.183, strong <=0.016.
Assessed · not applied · 11 not met · 8 not assessed
Pathogenic
PS1 Not met: no established pathogenic variant shares this p.Arg797His change - the only ClinVar record carrying it is Benign across 13 submissions.
PS2 Not assessed: no proband or parental genotypes exist in the record, so de novo status cannot be tested.
PS3 Not assessed: no functional assay of MLH3 p.Arg797His was found; OncoKB reports no variant-specific functional evidence for this variant.
PS4 Not met: no case-control or case-enrichment evidence exists, and gnomAD v4.1 grpmax AF 0.0647 (174 homozygotes) precludes enrichment in affected cases.
PM1 Not met: residue 797 lies outside MLH3's annotated ATPase (~1-349) and C-terminal MutL (~1189-1403) domains, is not a significant hotspot, and carries benign variation up to 6.61% popmax.
PM2 Not met: the allele is present at 0.35% overall in gnomAD v4.1 and 6.6% in African ancestry, far above the <=0.0001 PM2 threshold.
PM3 Not met: no pathogenic MLH3 variant in trans is reported, and this common allele (gnomAD v2.1 AF 0.64%, 72 homozygotes) is tolerated biallelically.
PM5 Not met: the other missense variants at codon 797 are uncertain or likely benign (p.Arg797Cys: 2 VUS + 1 likely benign; p.Arg797Asp: VUS), with none pathogenic.
PM6 Not assessed: no proband and no assumed-de-novo observation were recorded, despite 174 homozygotes in gnomAD.
PP1 Not assessed: no pedigree or relative genotypes exist, leaving zero informative meioses for a segregation test.
PP2 Not met: MLH3 is missense-tolerant (gnomAD v2.1 mis_z 0.74, oe_mis 0.92) and missense is not an established mechanism (1 of 2,176 ClinVar missense variants P/LP).
PP3 Not met: REVEL 0.083 is far below the >=0.644 supporting PP3 threshold for this missense variant.
PP4 Not assessed: the case supplies no proband phenotype or family history, and MLH3 is absent from the MMR gene sets that define a specific phenotype (PMID:34043773).
PP5 Not met: ClinVar VCV000314383 has 13 submissions from clinical laboratories and zero expert-panel classifications, none asserting pathogenic, so PP5's expert-panel prerequisite is absent.
Benign
BS3 Not assessed: no MLH3 p.Arg797His functional assay exists, and the REVEL 0.083 in-silico score is not BS3 experimental evidence.
BS4 Not assessed: no family genotypes exist, so non-segregation cannot be shown and 174 gnomAD homozygotes are not a BS4 finding.
BP2 Not met: no source places this variant in cis or in trans with a pathogenic allele; none of 13 ClinVar submissions reports phase.
BP5 Not assessed: no proband-level genotype data exist and no ClinVar submission documents a co-occurring alternate molecular cause for c.2390G>A.
BP6 Not met: the Benign ClinVar label on VCV000314383 is a 2-star aggregate of 12 laboratory submissions with zero expert-panel submissions, so BP6's expert-panel prerequisite is absent.
N/A · 5 PVS1 · PM4 · BS2 · BP3 · BP7
Research & evidence
Population frequency · supports benign
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
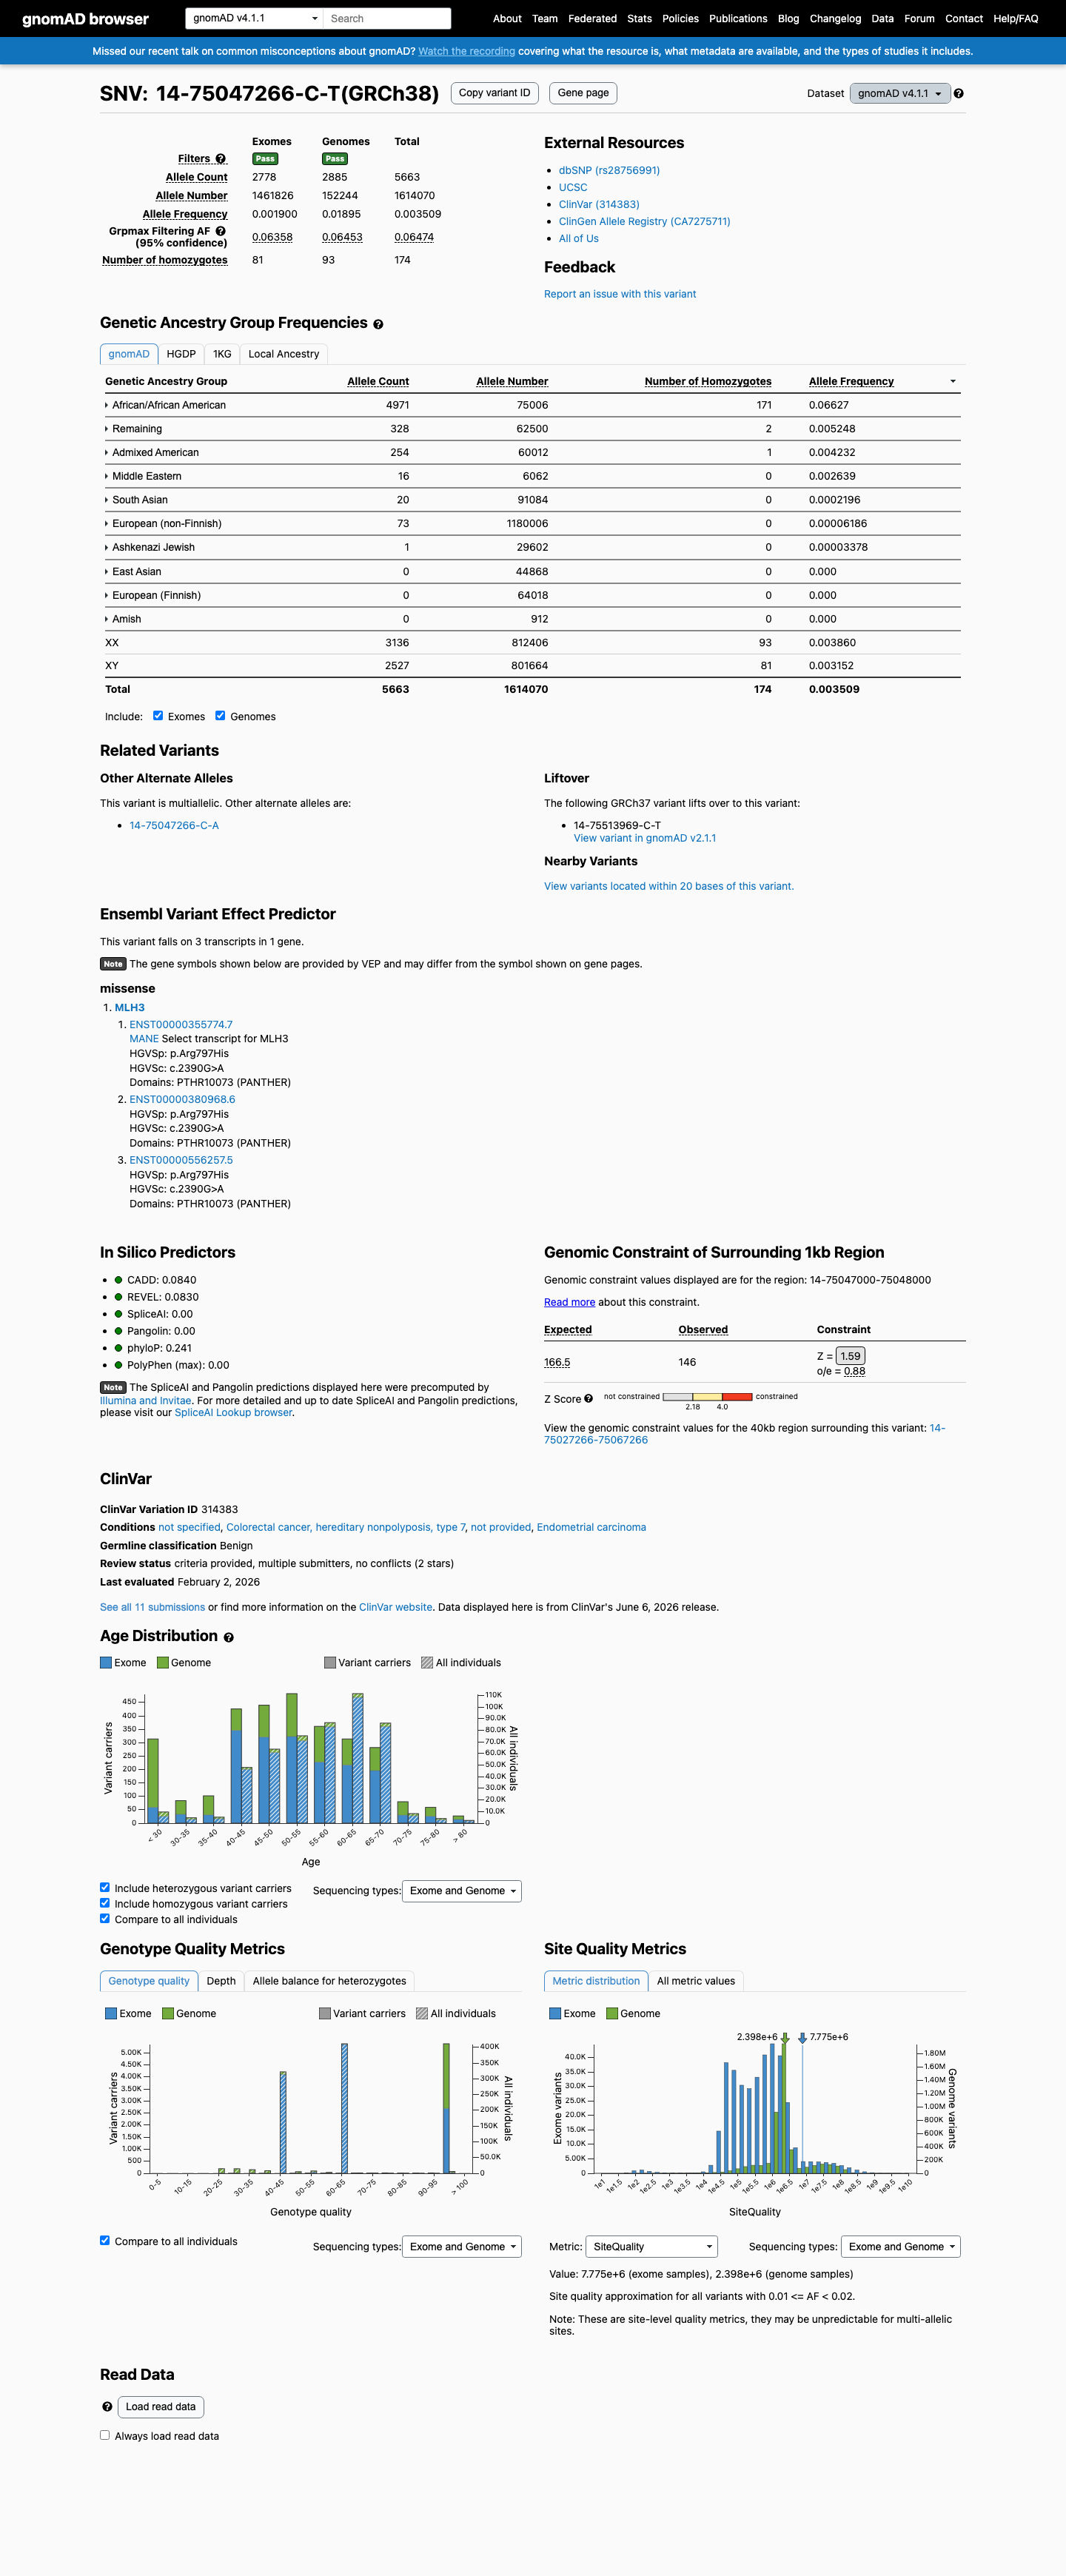
This variant is present in gnomAD v4.1 (AF= 0.00350852; MAF= 0.35085%, 5663/1614070 alleles, homozygotes = 174) and has highest observed frequency in the African/African American population (AF= 0.0662747; MAF= 6.62747%, 4971/75006 alleles, homozygotes = 171); grpmax FAF= 0.064736.
v2.1
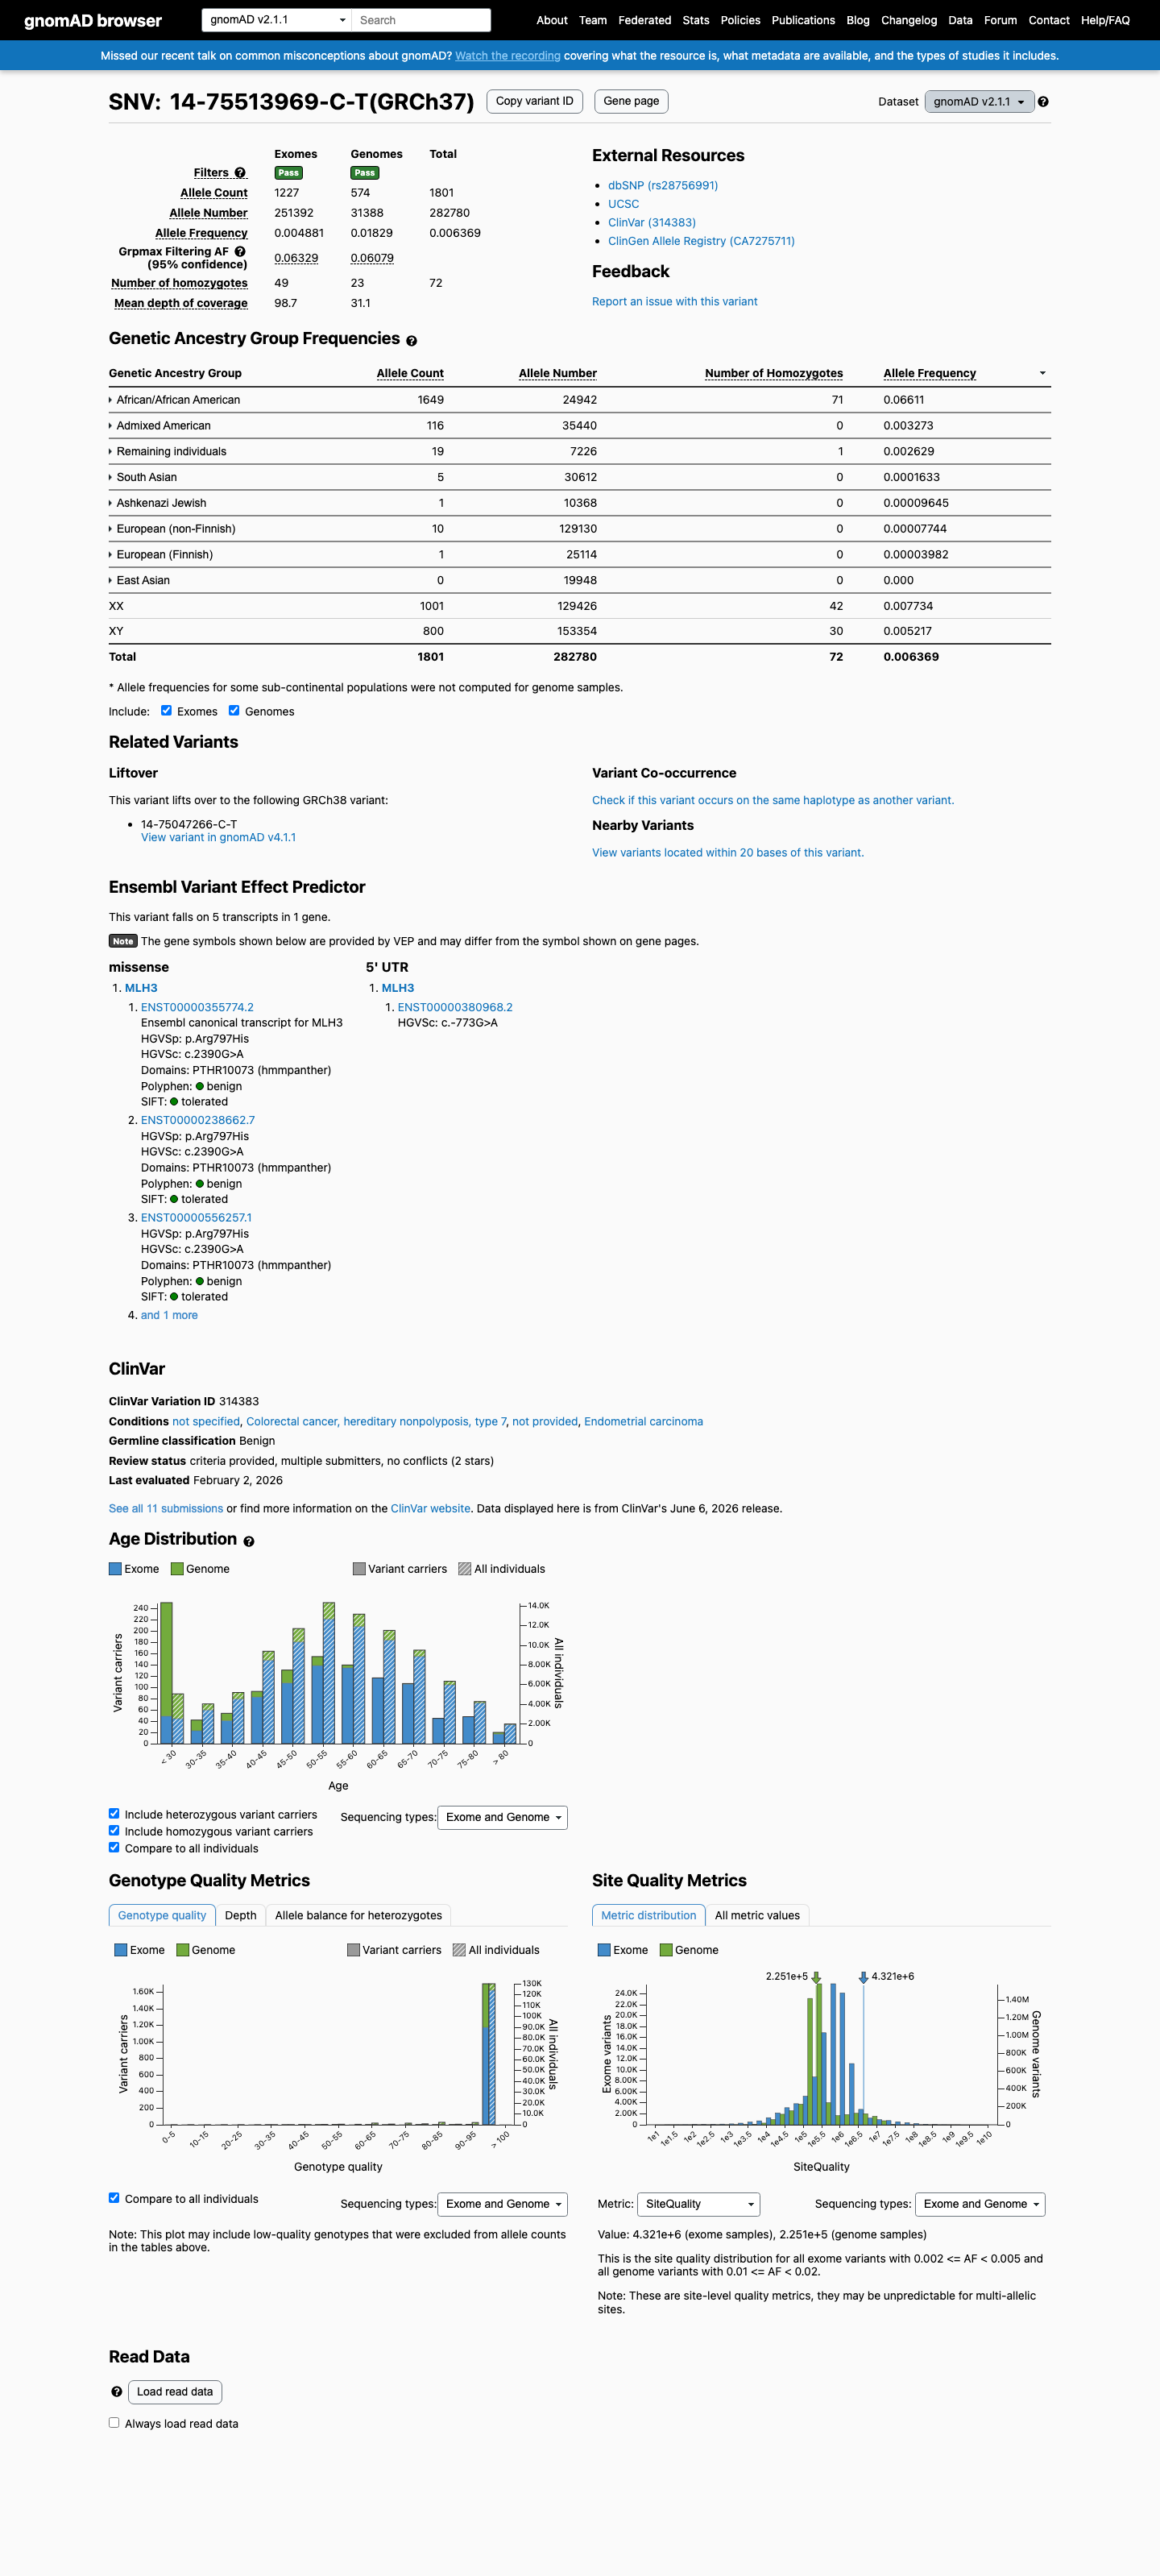
This variant is present in gnomAD v2.1 (AF= 0.00636891; MAF= 0.63689%, 1801/282780 alleles, homozygotes = 72) and has highest observed frequency in the African/African American population (AF= 0.0661134; MAF= 6.61134%, 1649/24942 alleles, homozygotes = 71); grpmax FAF= 0.0632924.
🇨🇦 CA
This variant is present in gnomAD-Canada v1.0 (AF= 0.005212858384013901, 96/18416 alleles, homozygotes = 5).
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.35% · 5663 / 1,614,070
174 hom · FAF 6.5%
African/African American
4971 / 75,006
6.6%
171 hom
Remaining individuals
328 / 62,500
0.52%
2 hom
Admixed American
254 / 60,012
0.42%
1 hom
Middle Eastern
16 / 6,062
0.26%
South Asian
20 / 91,084
0.022%
European (non-Finnish)
73 / 1,180,006
0.0062%
Ashkenazi Jewish
1 / 29,602
0.0034%
+ 3 not observed (European (Finnish), Amish, East Asian)
gnomAD v2.1
0.64% · 1801 / 282,780
72 hom · FAF 6.3%
African/African American
1649 / 24,942
6.6%
71 hom
Admixed American
116 / 35,440
0.33%
Remaining individuals
19 / 7,226
0.26%
1 hom
South Asian
5 / 30,612
0.016%
Ashkenazi Jewish
1 / 10,368
0.0096%
European (non-Finnish)
10 / 129,130
0.0077%
European (Finnish)
1 / 25,114
0.004%
+ 1 not observed (East Asian)
gnomAD Canada 🇨🇦
0.52% · 96 / 18,416
5 hom · FAF 7.1%
African/African American
87 / 1,020
8.5%
5 hom
Middle Eastern
1 / 144
0.69%
Remaining individuals
4 / 1,136
0.35%
Latino/Admixed American
2 / 838
0.24%
European (non-Finnish)
2 / 11,738
0.017%
+ 4 not observed (Ashkenazi Jewish, East Asian, European (Finnish), South Asian)
ClinVar screenshot
ClinVar
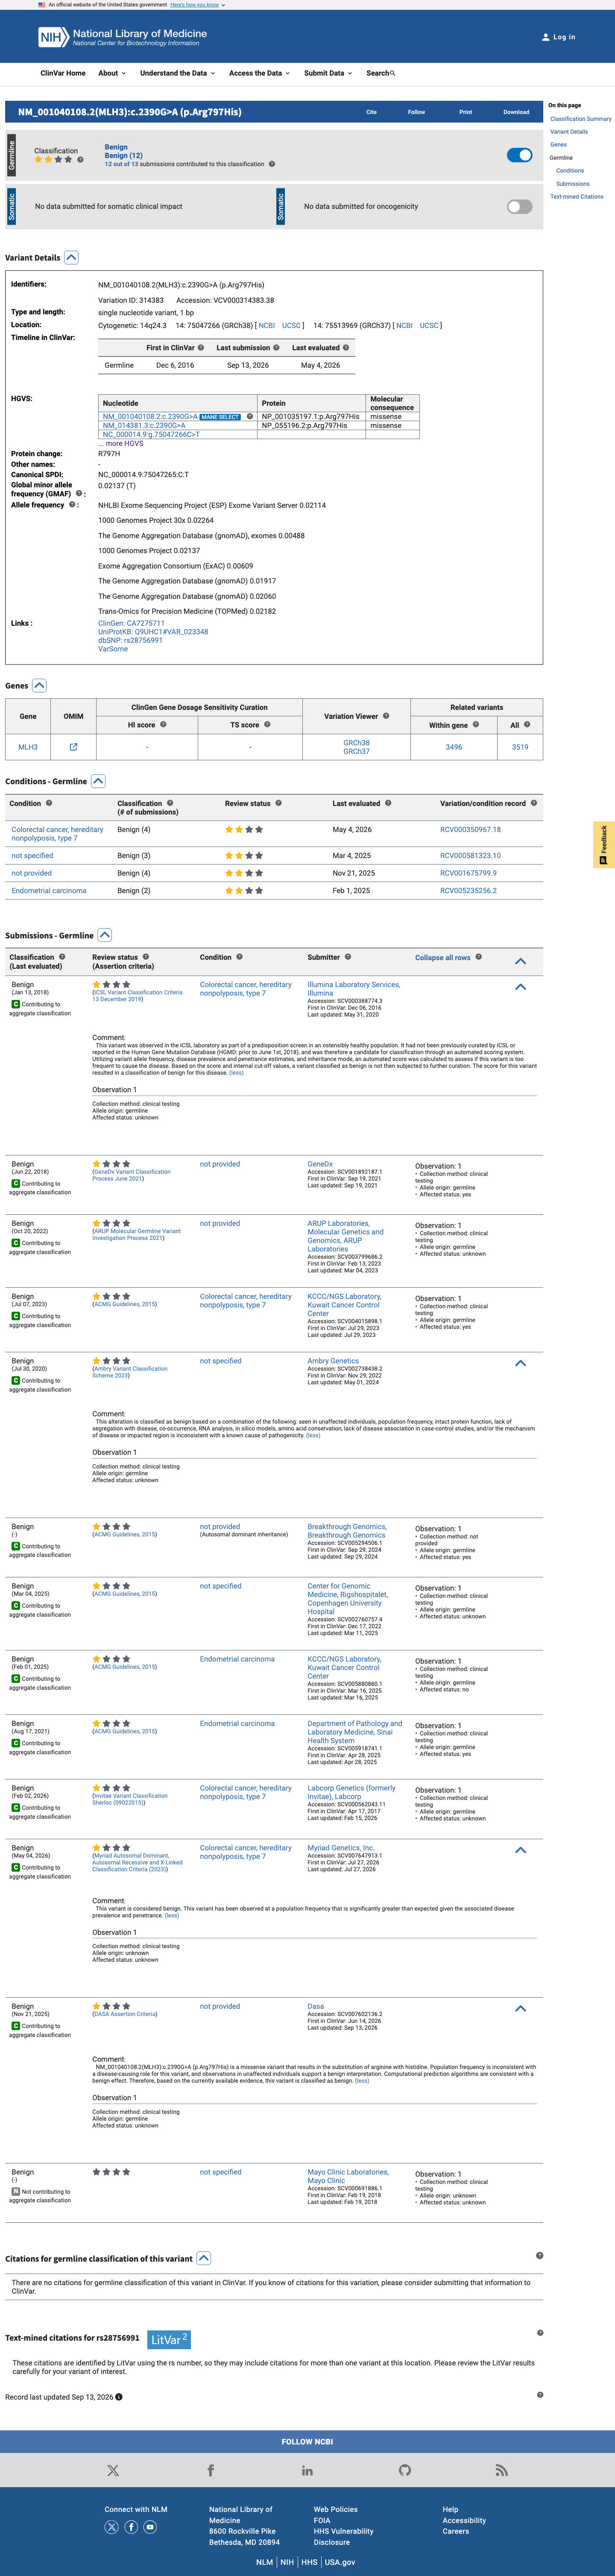
This variant has been reported in ClinVar as Benign (12 clinical laboratories). (ClinVarID = 314383)
SpliceAI screenshot
In silico
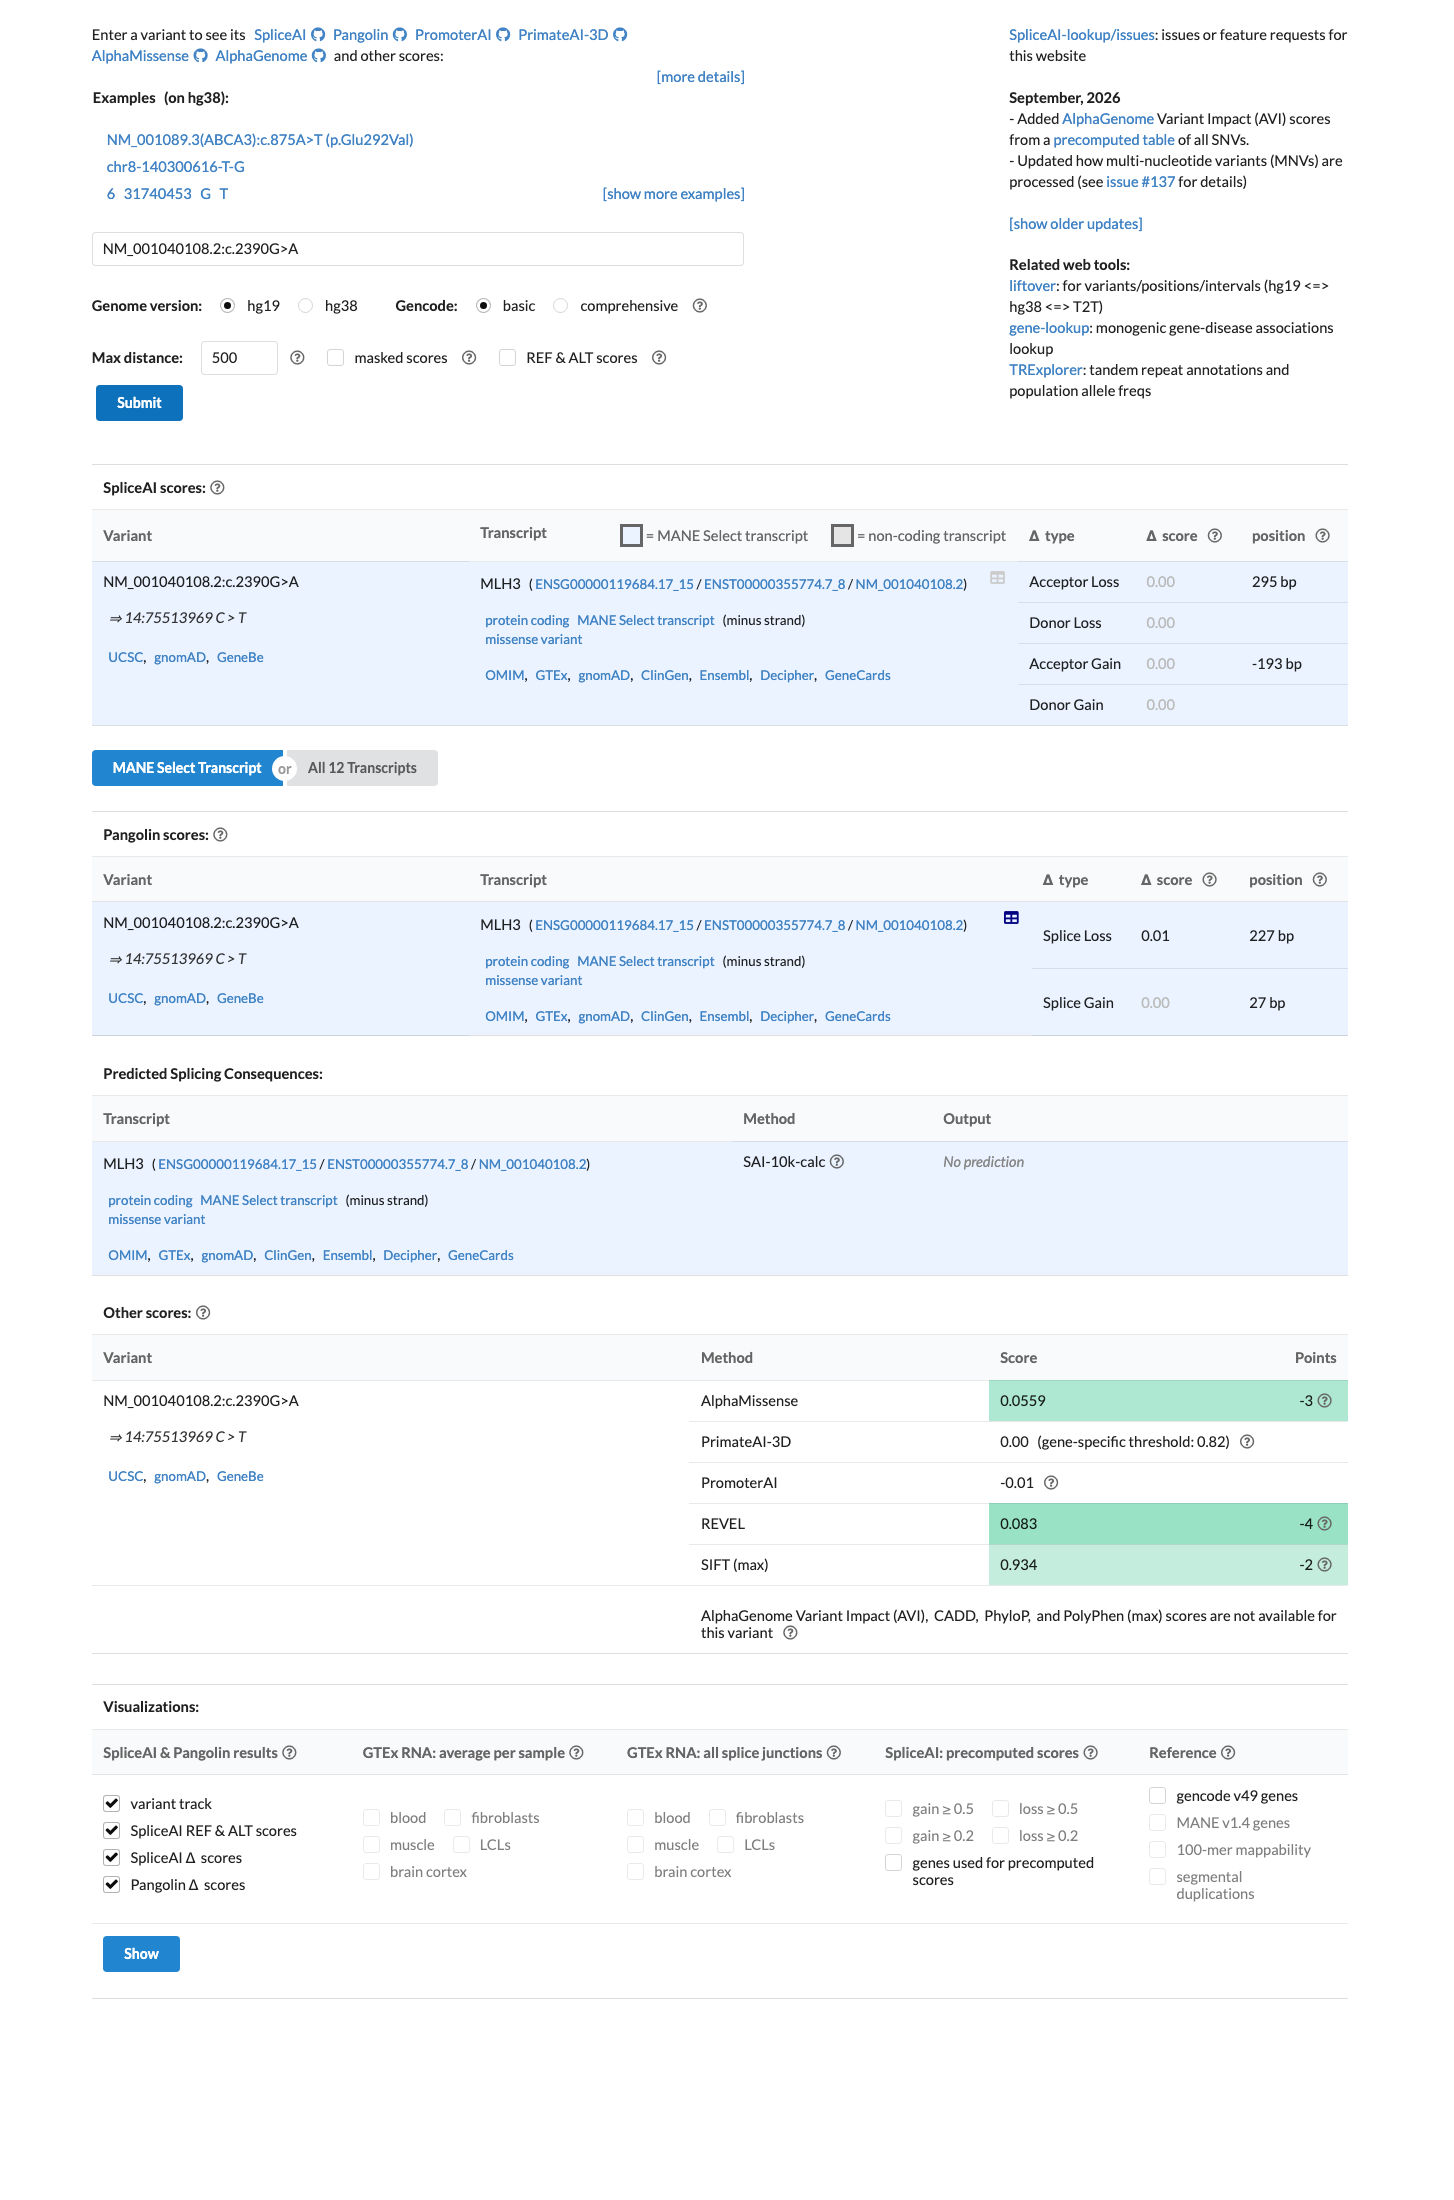
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.00). REVEL score = 0.083. BayesDel score = -0.564808.
Functional / OncoKB screenshot
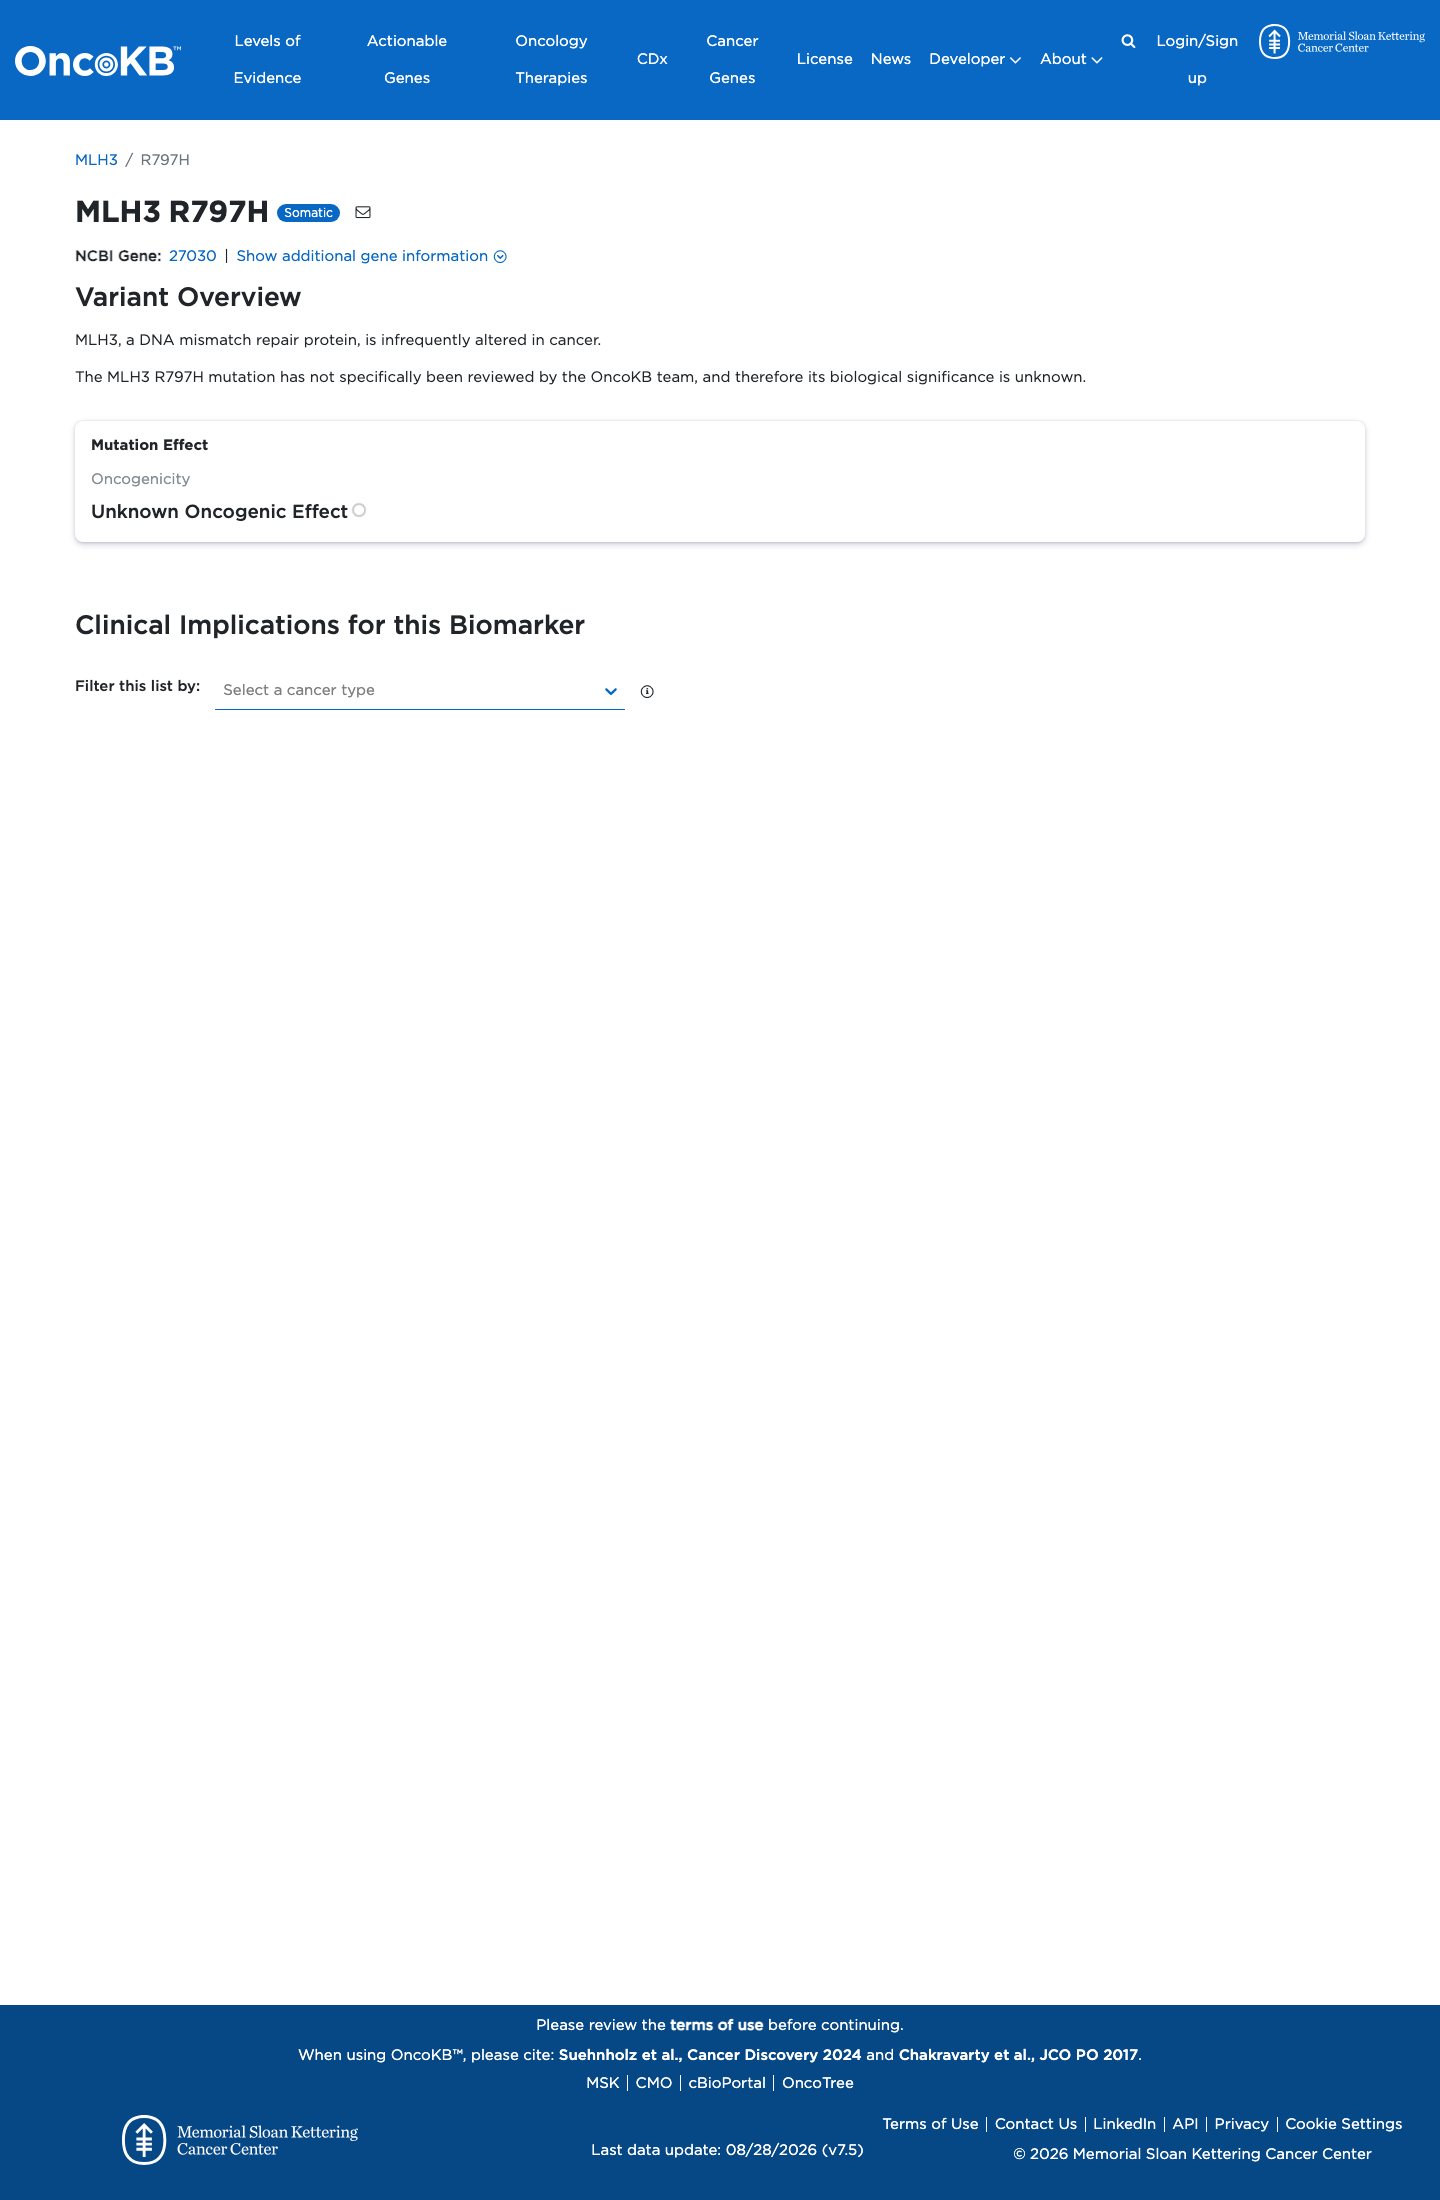
Functional Unknown Oncogenic Effect
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. MLH3, a DNA mismatch repair protein, is infrequently altered in cancer.
OncoKB ↗
COSMIC screenshot
COSMIC
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
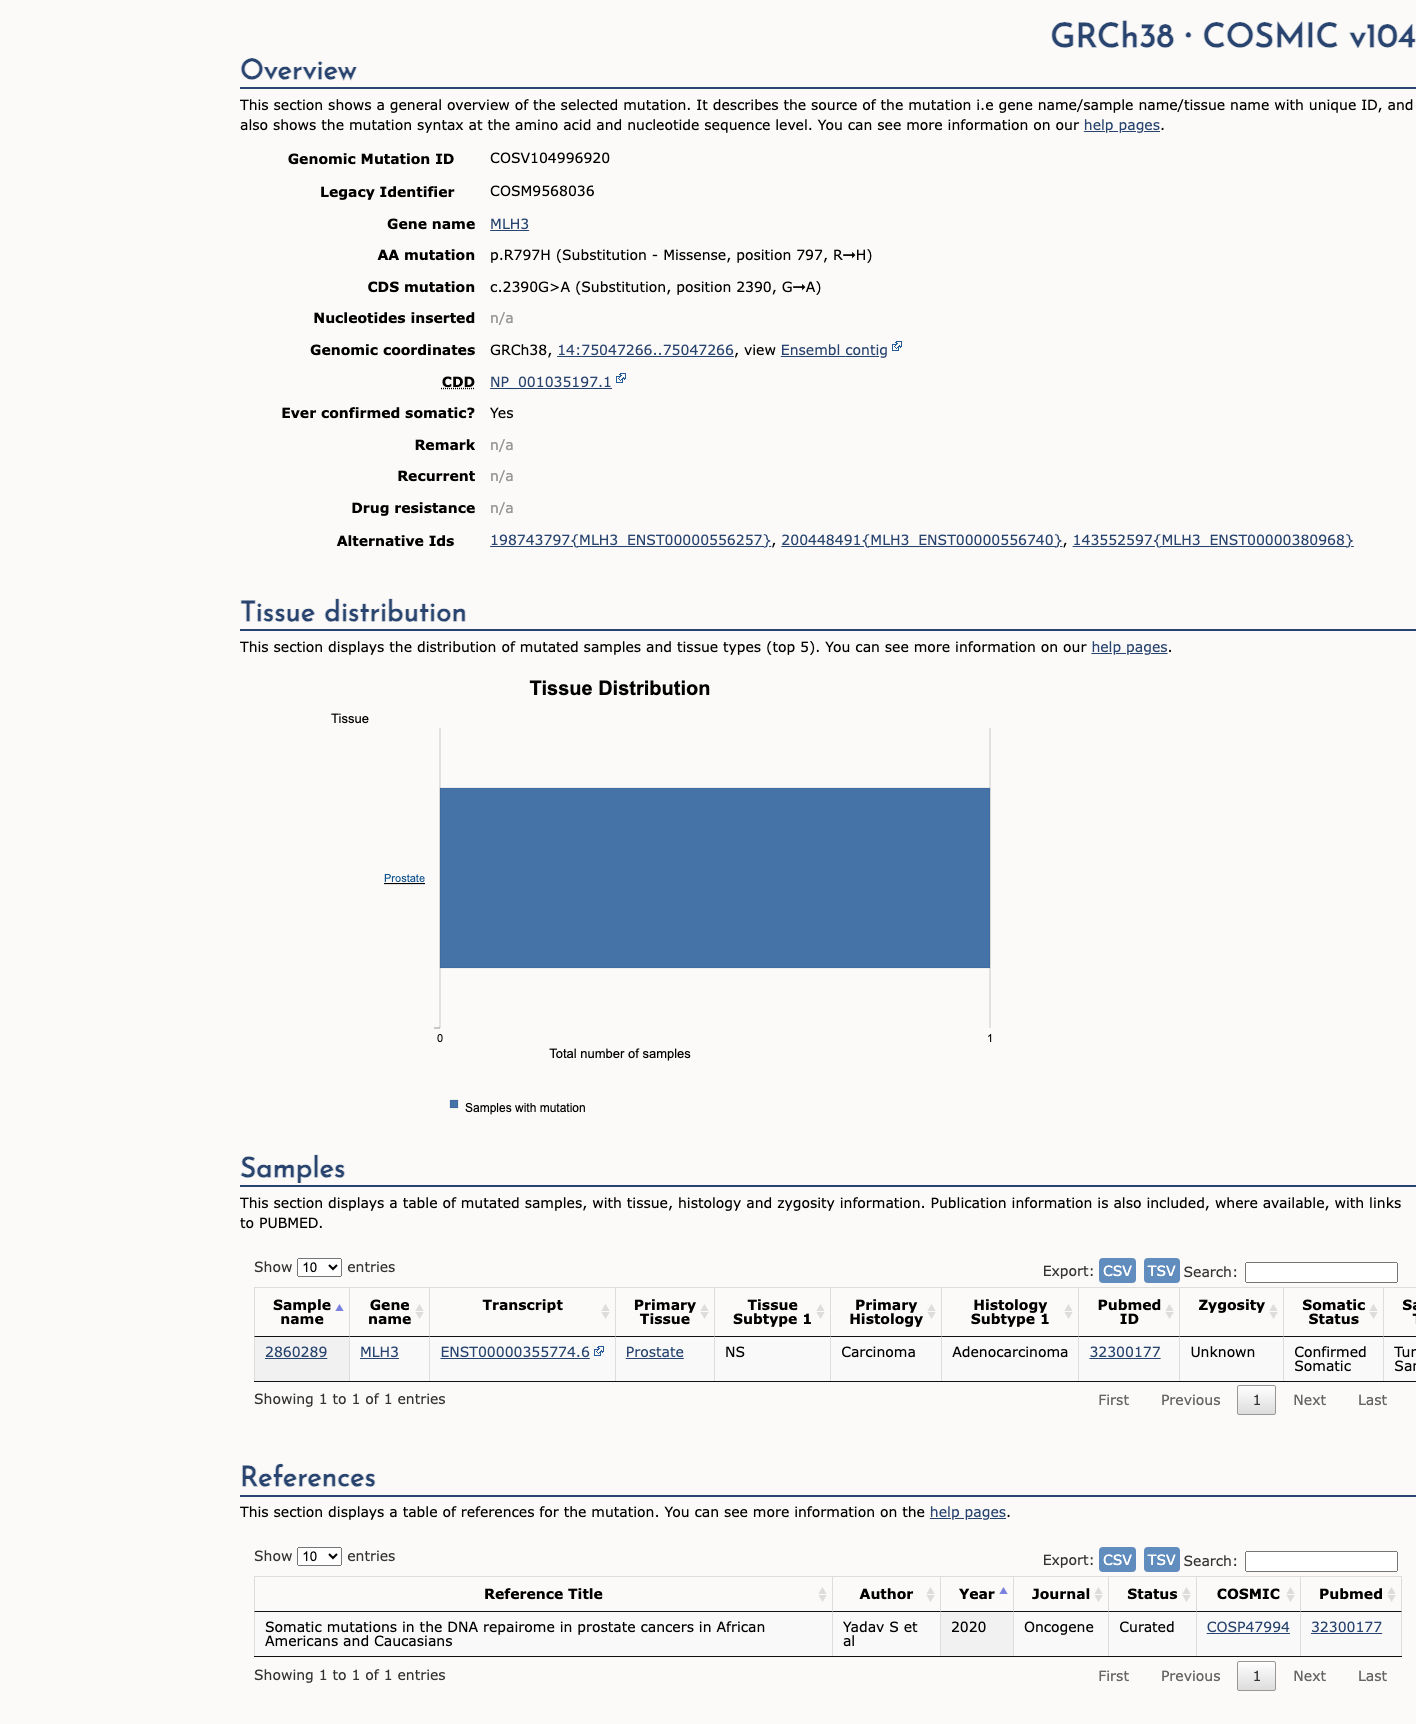
This variant does not lie in a statistically significant hotspot. This variant has previously been reported in somatic cancers (COSMIC; COSV104996920, n = 1 times).
Hotspots
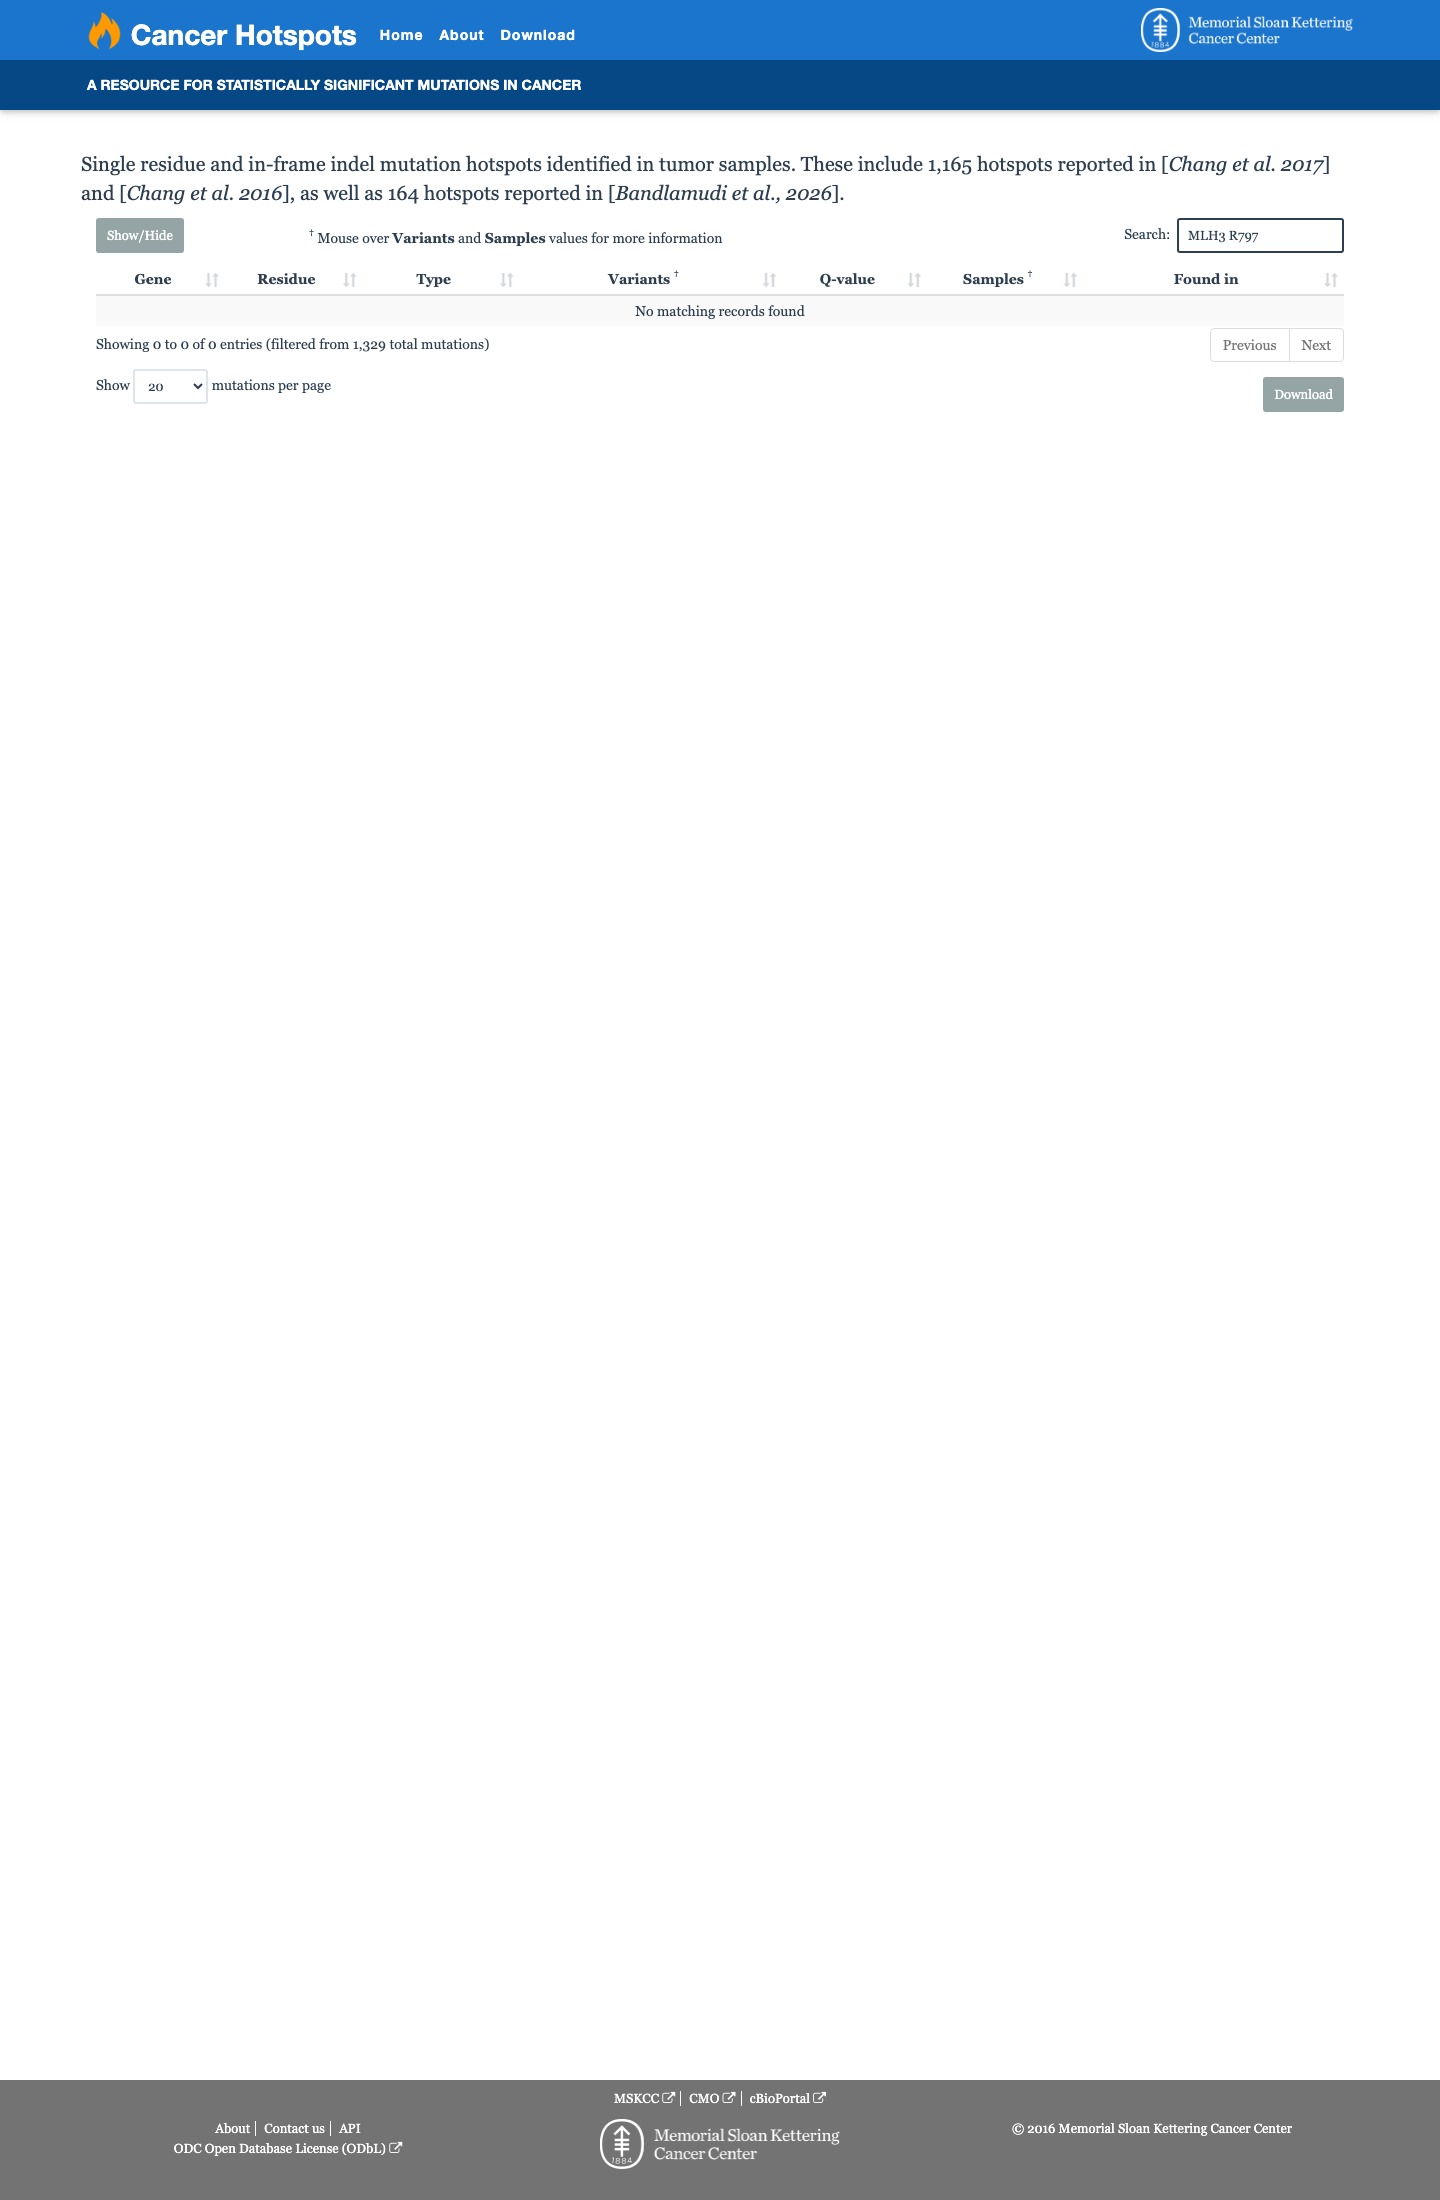
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
4papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 3 further PMIDs triaged but not cited — see Sources & references.
Rule & framework references · cited for criterion definitions, not variant evidence
24493721 ↗ American Society of Clinical Oncology Expert Statement: collection and use of a cancer family history for oncology providers.
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.
33516529 ↗ Endometrial cancer: A society of gynecologic oncology evidence-based review and recommendations.
34043773 ↗ European guidelines from the EHTG and ESCP for Lynch syndrome: an updated third edition of the Mallorca guidelines based on gene and gender.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 3 PMIDs not cited in assessment
33451724 ↗ Endometrial cancer: A society of gynecologic oncology evidence-based review and recommendations, part II. CLINVAR
24905773 ↗ Endometrial cancer: a review and current management strategies: part I. CLINVAR
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification criteria. CLINVAR