Back
NM_001042492.2:c.7584A>G
p.Gln2528= · NF1
0%
complete
Final classification
Likely Benign
BS2BP4BP7
NF1
c.7584A>G
p.Gln2528=
synonymous · exon 51

NF1 encodes neurofibromin, a tumor suppressor protein that negatively regulates the RAS signal transduction pathway. As a GTPase-activating protein, it helps switch RAS proteins from their active to inactive state, keeping cell growth in check; loss of NF1 function leaves RAS overactive and drives downstream growth pathways such as MAPK/ERK and PI3K. Inherited changes in NF1 cause the cancer-predisposition syndrome neurofibromatosis type 1, and are also linked to juvenile myelomonocytic leukemia and Watson syndrome. Somatic changes in NF1 are found in many tumor types, including breast cancer, melanoma, and glioma.

This variant

NF1 encodes neurofibromin, a RAS-pathway tumor suppressor, and inherited NF1 variants cause neurofibromatosis type 1 through loss of growth control.

Transcript
NM_001042492.2
HGVS · transcript:coding
NM_001042492.2:c.7584A>G
GRCh38
chr17:31352383 A>G
GRCh37
chr17:29679401 A>G
Likely Benign: BS2 (strong), BP4 (supporting), and BP7 (supporting) satisfy the generic ACMG/AMP fallback combination of one strong plus one supporting benign criterion.
Classification rationale
BS2BP4BP7 Likely Benign
NF1 c.7584A>G synonymous · exon 51

BS2 strong: gnomAD v4.1 reports six homozygotes, inconsistent with a highly penetrant autosomal-dominant NF1 allele. BP4 supporting: SpliceAI maximum delta 0.001 predicts no splice impact. BP7 supporting: the synonymous variant is outside the canonical splice consensus with minimal predicted splice impact.

BS2 + BP4 + BP7 → Likely Benign
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_001042492.2 · variants mapped to exon structure
NF1 NM_001042492.2
Fetching transcript structure from UCSC…
Applied criteria · 3 applied · 16 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 3
Strength Supporting Moderate Strong Very strong
BS2 strong review Benign
Met: gnomAD v4.1 contains 6 homozygotes, incompatible with a highly penetrant autosomal-dominant NF1 allele.
The governing NF1 framework identifies autosomal-dominant inheritance for neurofibromatosis type 1.gnomAD v4.1 reports total_hom 6, comprising 5 exome homozygotes and 1 genome homozygote, among 1578/1614034 observed alleles.The absence of individual-level phenotype and age data prevents direct confirmation of the ACMG/AMP healthy-adult wording and is the reason for human review.
BP4 supporting Benign
Met, supporting: synonymous variant has a SpliceAI maximum delta score of 0.001, meeting the <=0.1 BP4 threshold.
The case variant is NM_001042492.2:c.7584A>G, annotated as synonymous, NP_001035957.1:p.(Gln2528=).SpliceAI is available and reports maximum delta 0.001.The supplied Jaganathan et al. 2019 calibration (PMID:30661751) defines BP4 supporting for SpliceAI max delta <=0.1.
BP7 supporting Benign
Met, supporting: synonymous c.7584A>G is non-canonical and has a minimal SpliceAI maximum delta score of 0.001.
The variant is synonymous, NM_001042492.2:c.7584A>G, p.(Gln2528=).The case assessment identifies the variant as outside the canonical splice consensus, and SpliceAI reports maximum delta 0.001.The ACMG/AMP guideline basis for BP7 is Richards et al. 2015 (PMID:25741868): a synonymous variant may support benign evidence when it is not expected to affect splicing.
Assessed · not applied · 7 not met · 9 not assessed
Pathogenic
PS2 Not assessed: no confirmed parental genotypes or maternity/paternity evidence establish that c.7584A>G arose de novo in the proband.
PS3 Not assessed: no variant-specific validated functional assay or transcript result was reported for c.7584A>G (p.Gln2528=).
PS4 Not met: reviewed studies provide no exact-variant affected-case series or case-control enrichment for NF1 c.7584A>G.
PM2 Not met: gnomAD v4.1 overall AF is 0.000977675, above the 0.0001 supporting PM2 threshold.
PM6 Not assessed: no documented family testing supports even a presumed de novo origin of c.7584A>G.
PP1 Not assessed: zero informative affected-relative genotypes or meioses are documented for segregation of c.7584A>G.
PP3 Not met: synonymous variant uses SpliceAI, whose maximum delta score is 0.001, below the 0.2 PP3 supporting threshold.
PP4 Not assessed: no patient phenotype or disease-specific diagnostic features are documented for evaluating phenotype specificity.
PP5 Not met: ClinVar reports zero expert-panel submissions for the exact variant, and available laboratory labels are not eligible for PP5.
Benign
BA1 Not met: maximum observed all-comers population AF is 0.00909, below the 0.05 stand-alone threshold.
BS1 Not met: maximum observed all-comers population AF is 0.00909, below the 0.01 strong threshold.
BS3 Not assessed: no variant-specific validated assay demonstrated normal function or normal transcript processing for c.7584A>G (p.Gln2528=).
BS4 Not assessed: no informative unaffected relatives carrying c.7584A>G are documented for non-segregation.
BP2 Not assessed: no affected-proband observation with a second pathogenic variant and no reliable cis/trans phase information are documented.
BP5 Not assessed: no patient-level alternative molecular diagnosis is documented to explain the phenotype independently of this NF1 variant.
BP6 Not met: ClinVar has no exact-variant expert-panel classification, so laboratory Benign or Likely benign labels cannot trigger BP6.
N/A · 9 PVS1 · PS1 · PM1 · PM3 · PM4 · PM5 · PP2 · BP1 · BP3
Research & evidence
Population frequency · supports benign
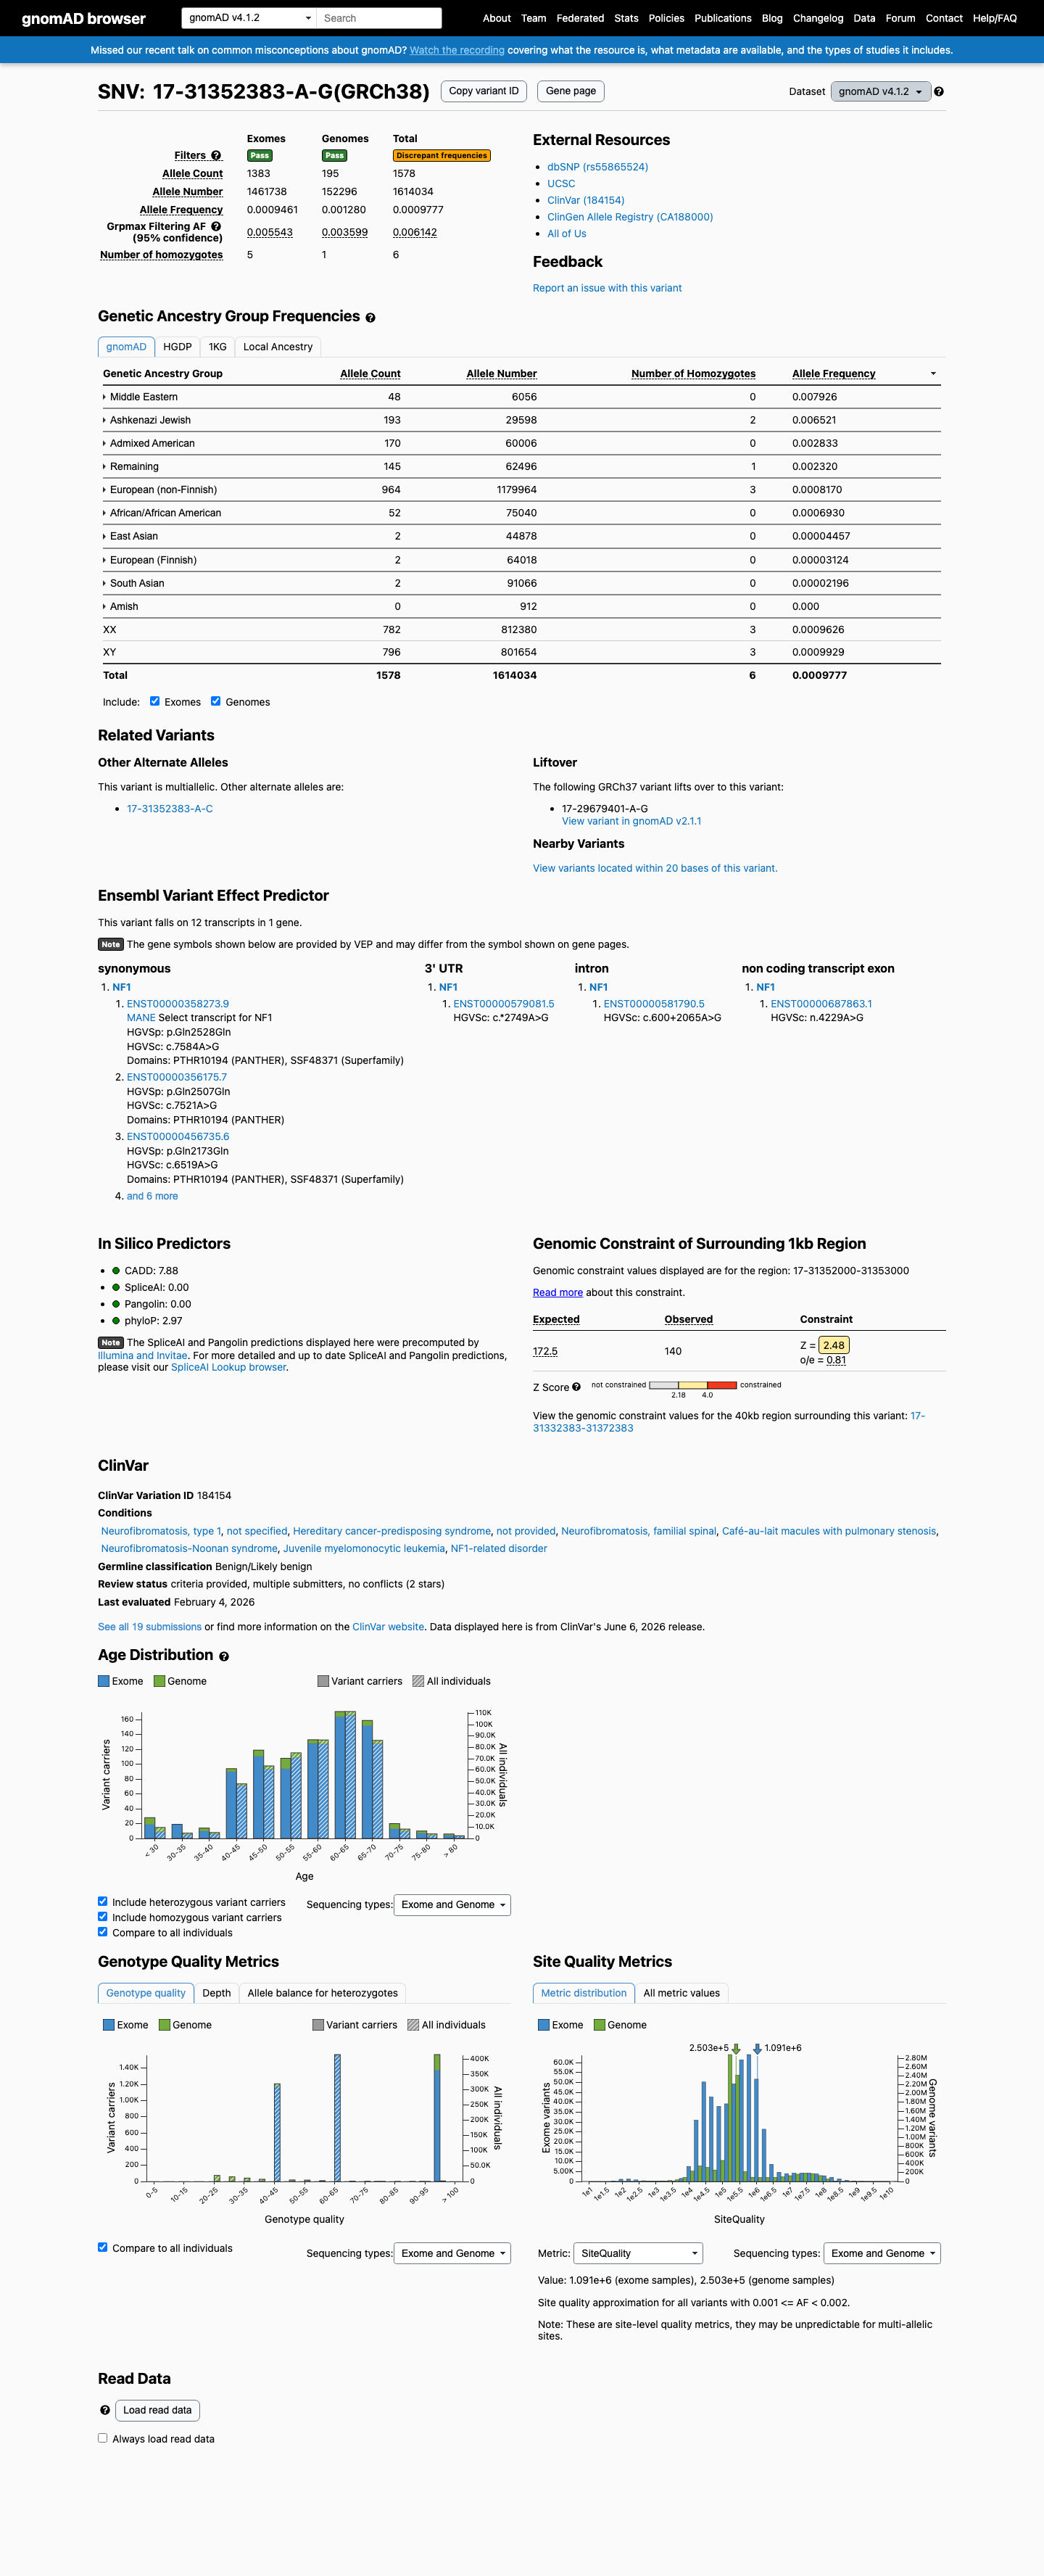
gnomAD v4.1 screenshot
gnomAD v4.1
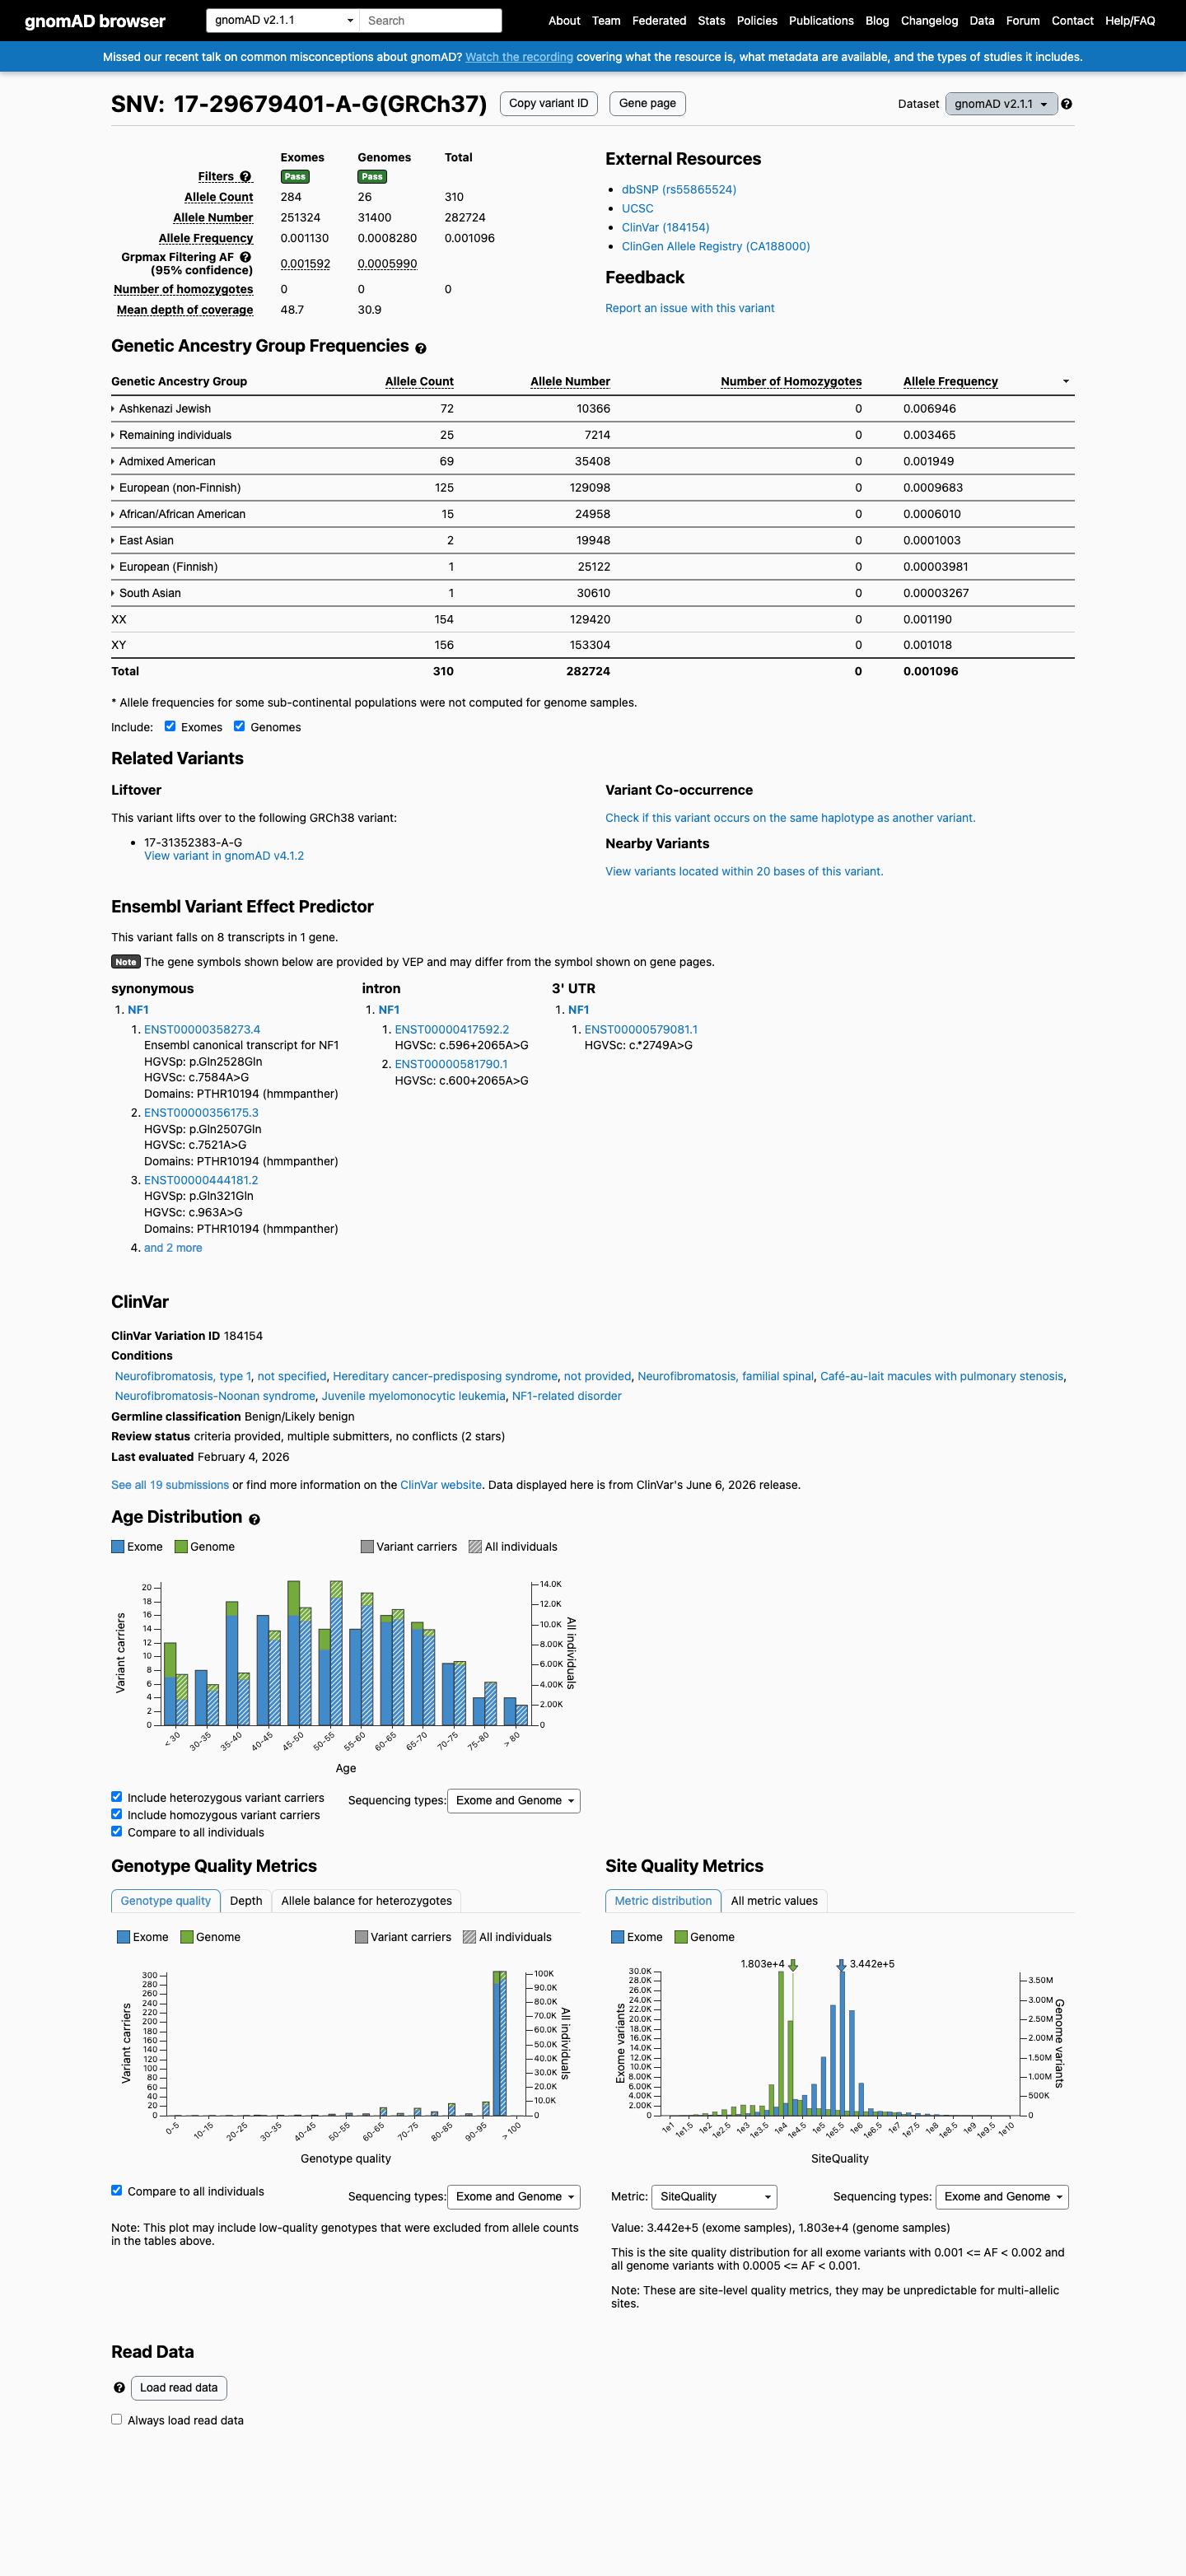
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 0.000977675; MAF= 0.09777%, 1578/1614034 alleles, homozygotes = 6) and has highest observed frequency in the Middle Eastern population (AF= 0.00792602; MAF= 0.79260%, 48/6056 alleles, homozygotes = 0); grpmax FAF= 0.0061424.
v2.1
This variant is present in gnomAD v2.1 (AF= 0.00109648; MAF= 0.10965%, 310/282724 alleles, homozygotes = 0) and has highest observed frequency in the Ashkenazi Jewish population (AF= 0.00694578; MAF= 0.69458%, 72/10366 alleles, homozygotes = 0); grpmax FAF= 0.00159181.
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.098% · 1578 / 1,614,034
6 hom · FAF 0.61%
Middle Eastern
48 / 6,056
0.79%
Ashkenazi Jewish
193 / 29,598
0.65%
2 hom
Admixed American
170 / 60,006
0.28%
Remaining individuals
145 / 62,496
0.23%
1 hom
European (non-Finnish)
964 / 1,179,964
0.082%
3 hom
African/African American
52 / 75,040
0.069%
East Asian
2 / 44,878
0.0045%
European (Finnish)
2 / 64,018
0.0031%
South Asian
2 / 91,066
0.0022%
+ 1 not observed (Amish)
gnomAD v2.1
0.11% · 310 / 282,724
0 hom · FAF 0.16%
Ashkenazi Jewish
72 / 10,366
0.69%
Remaining individuals
25 / 7,214
0.35%
Admixed American
69 / 35,408
0.19%
European (non-Finnish)
125 / 129,098
0.097%
African/African American
15 / 24,958
0.06%
East Asian
2 / 19,948
0.01%
European (Finnish)
1 / 25,122
0.004%
South Asian
1 / 30,610
0.0033%
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
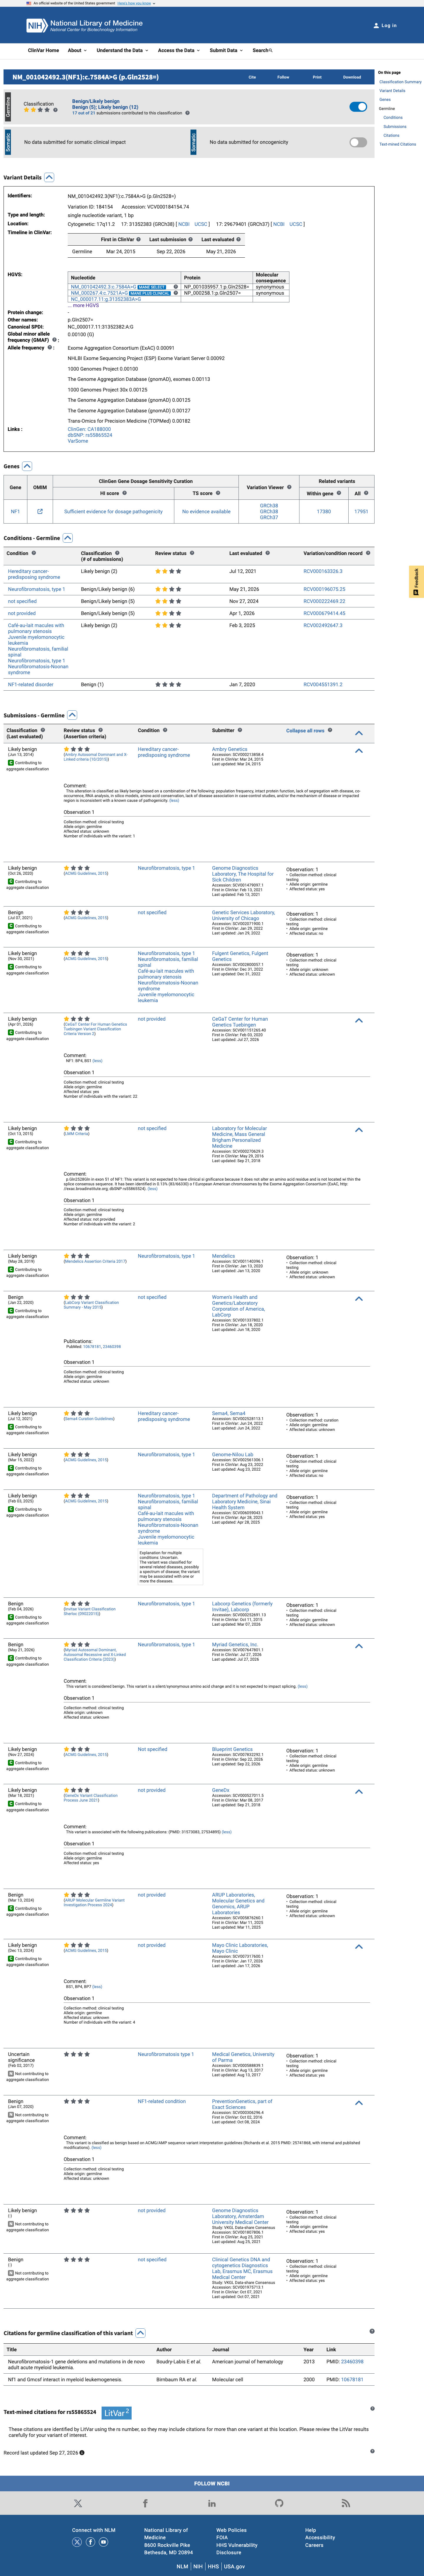
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Likely benign (12 clinical laboratories) and as Benign (7 clinical laboratories) and as Uncertain significance (1 clinical laboratory). (ClinVarID = 184154)
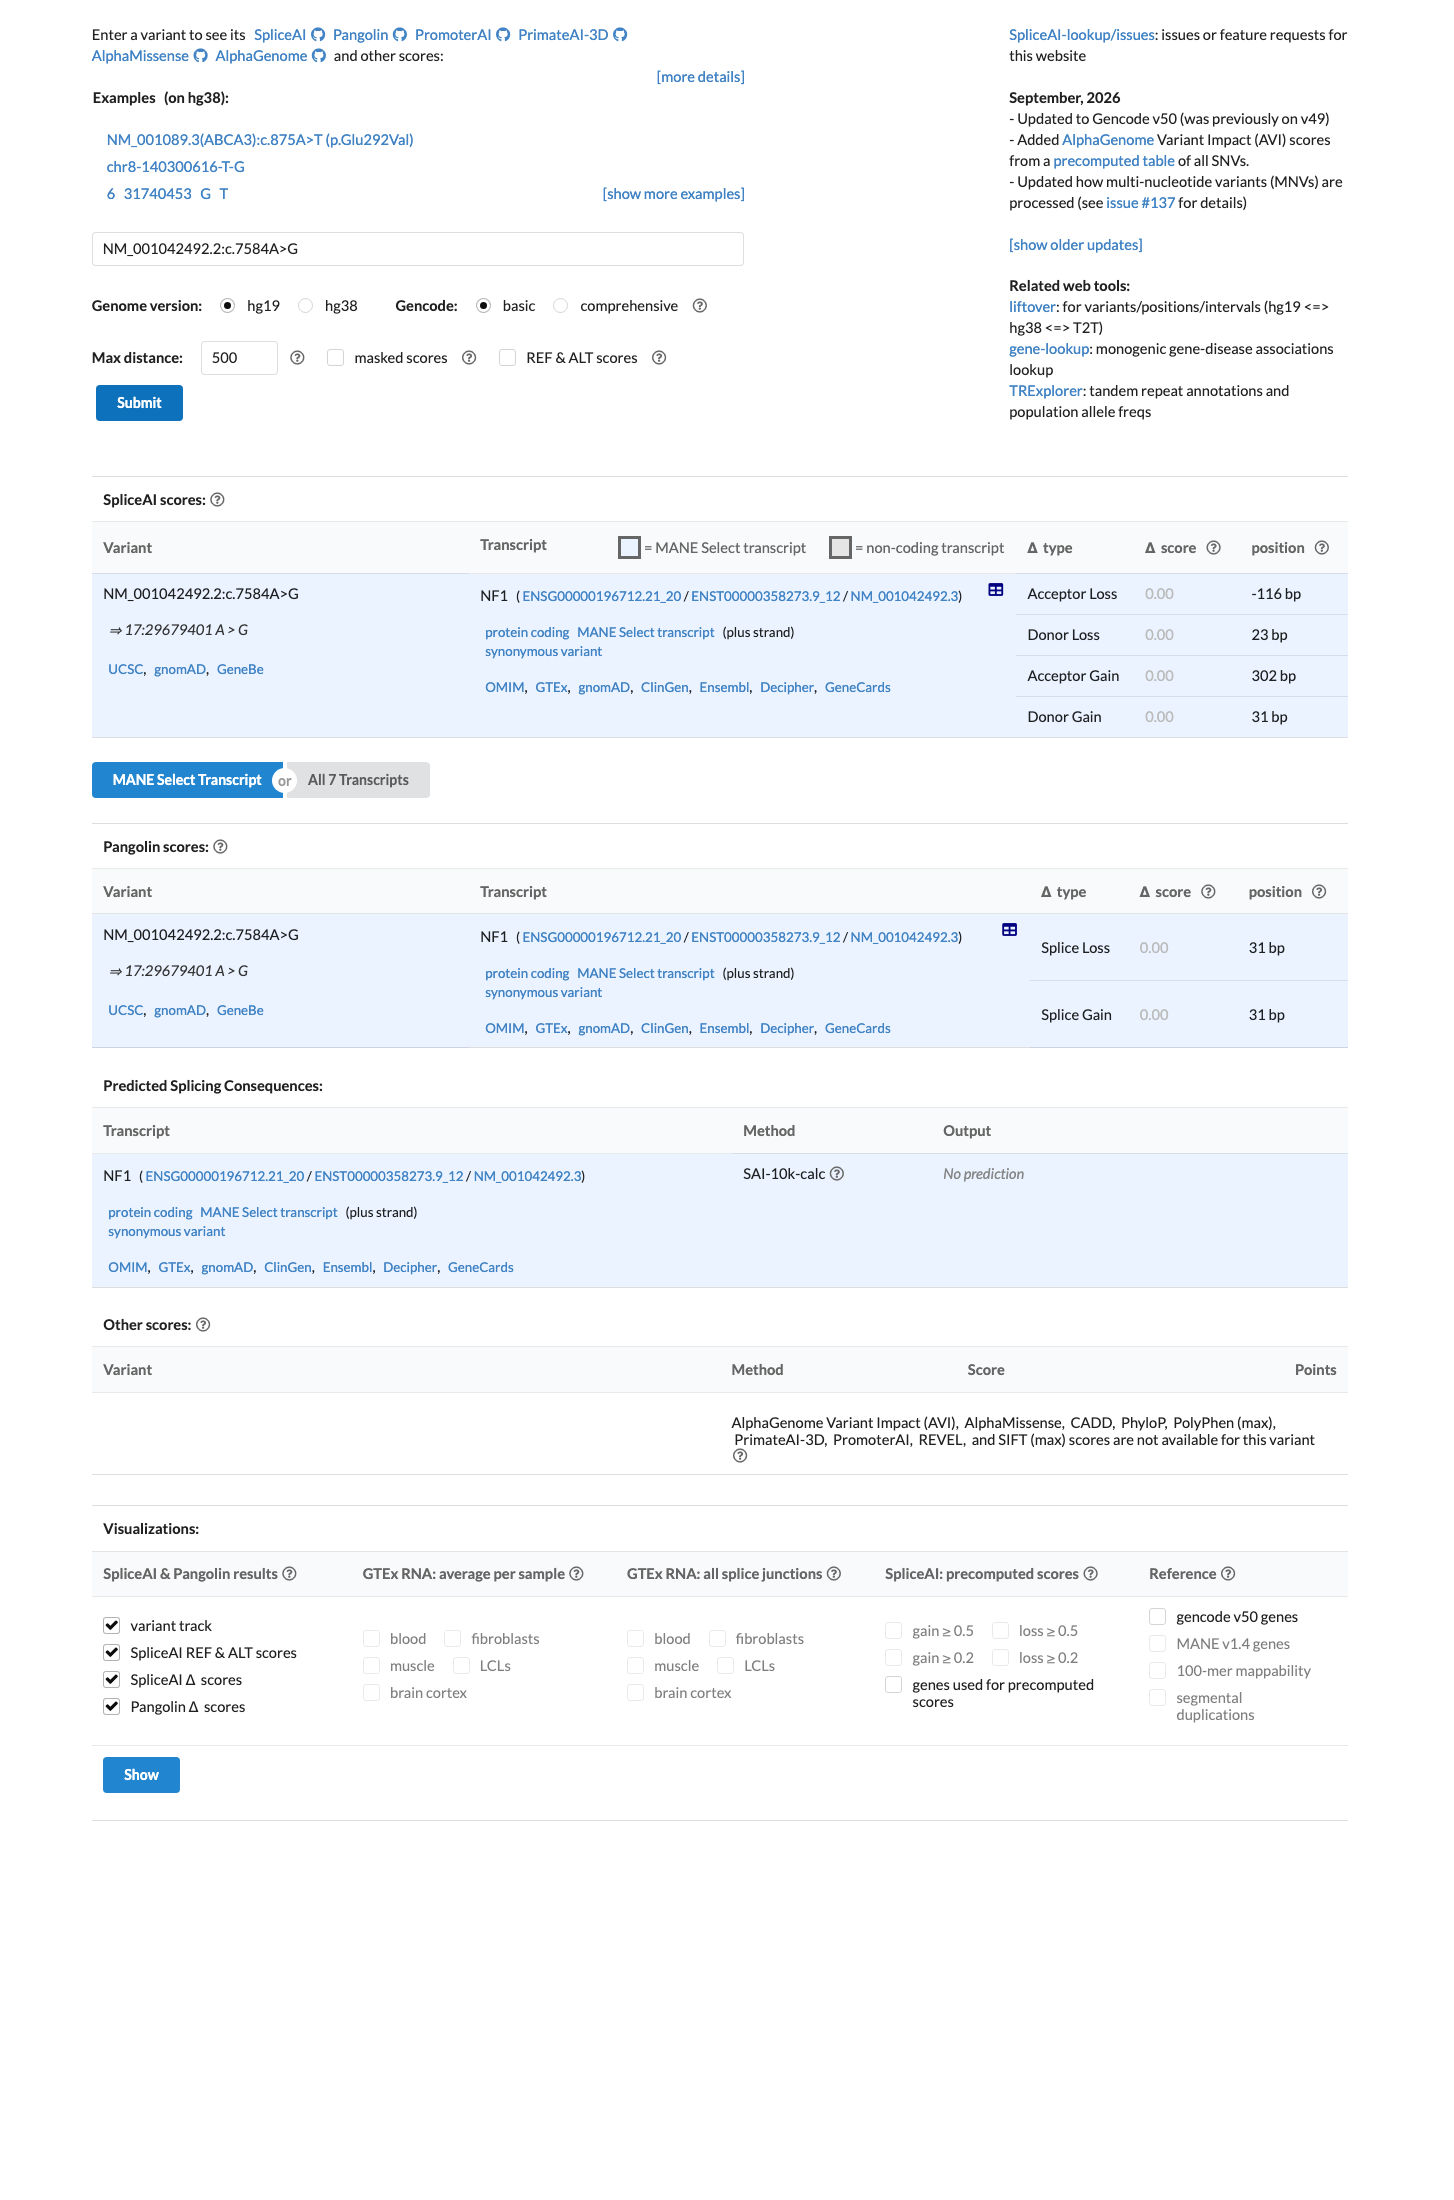
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.00).
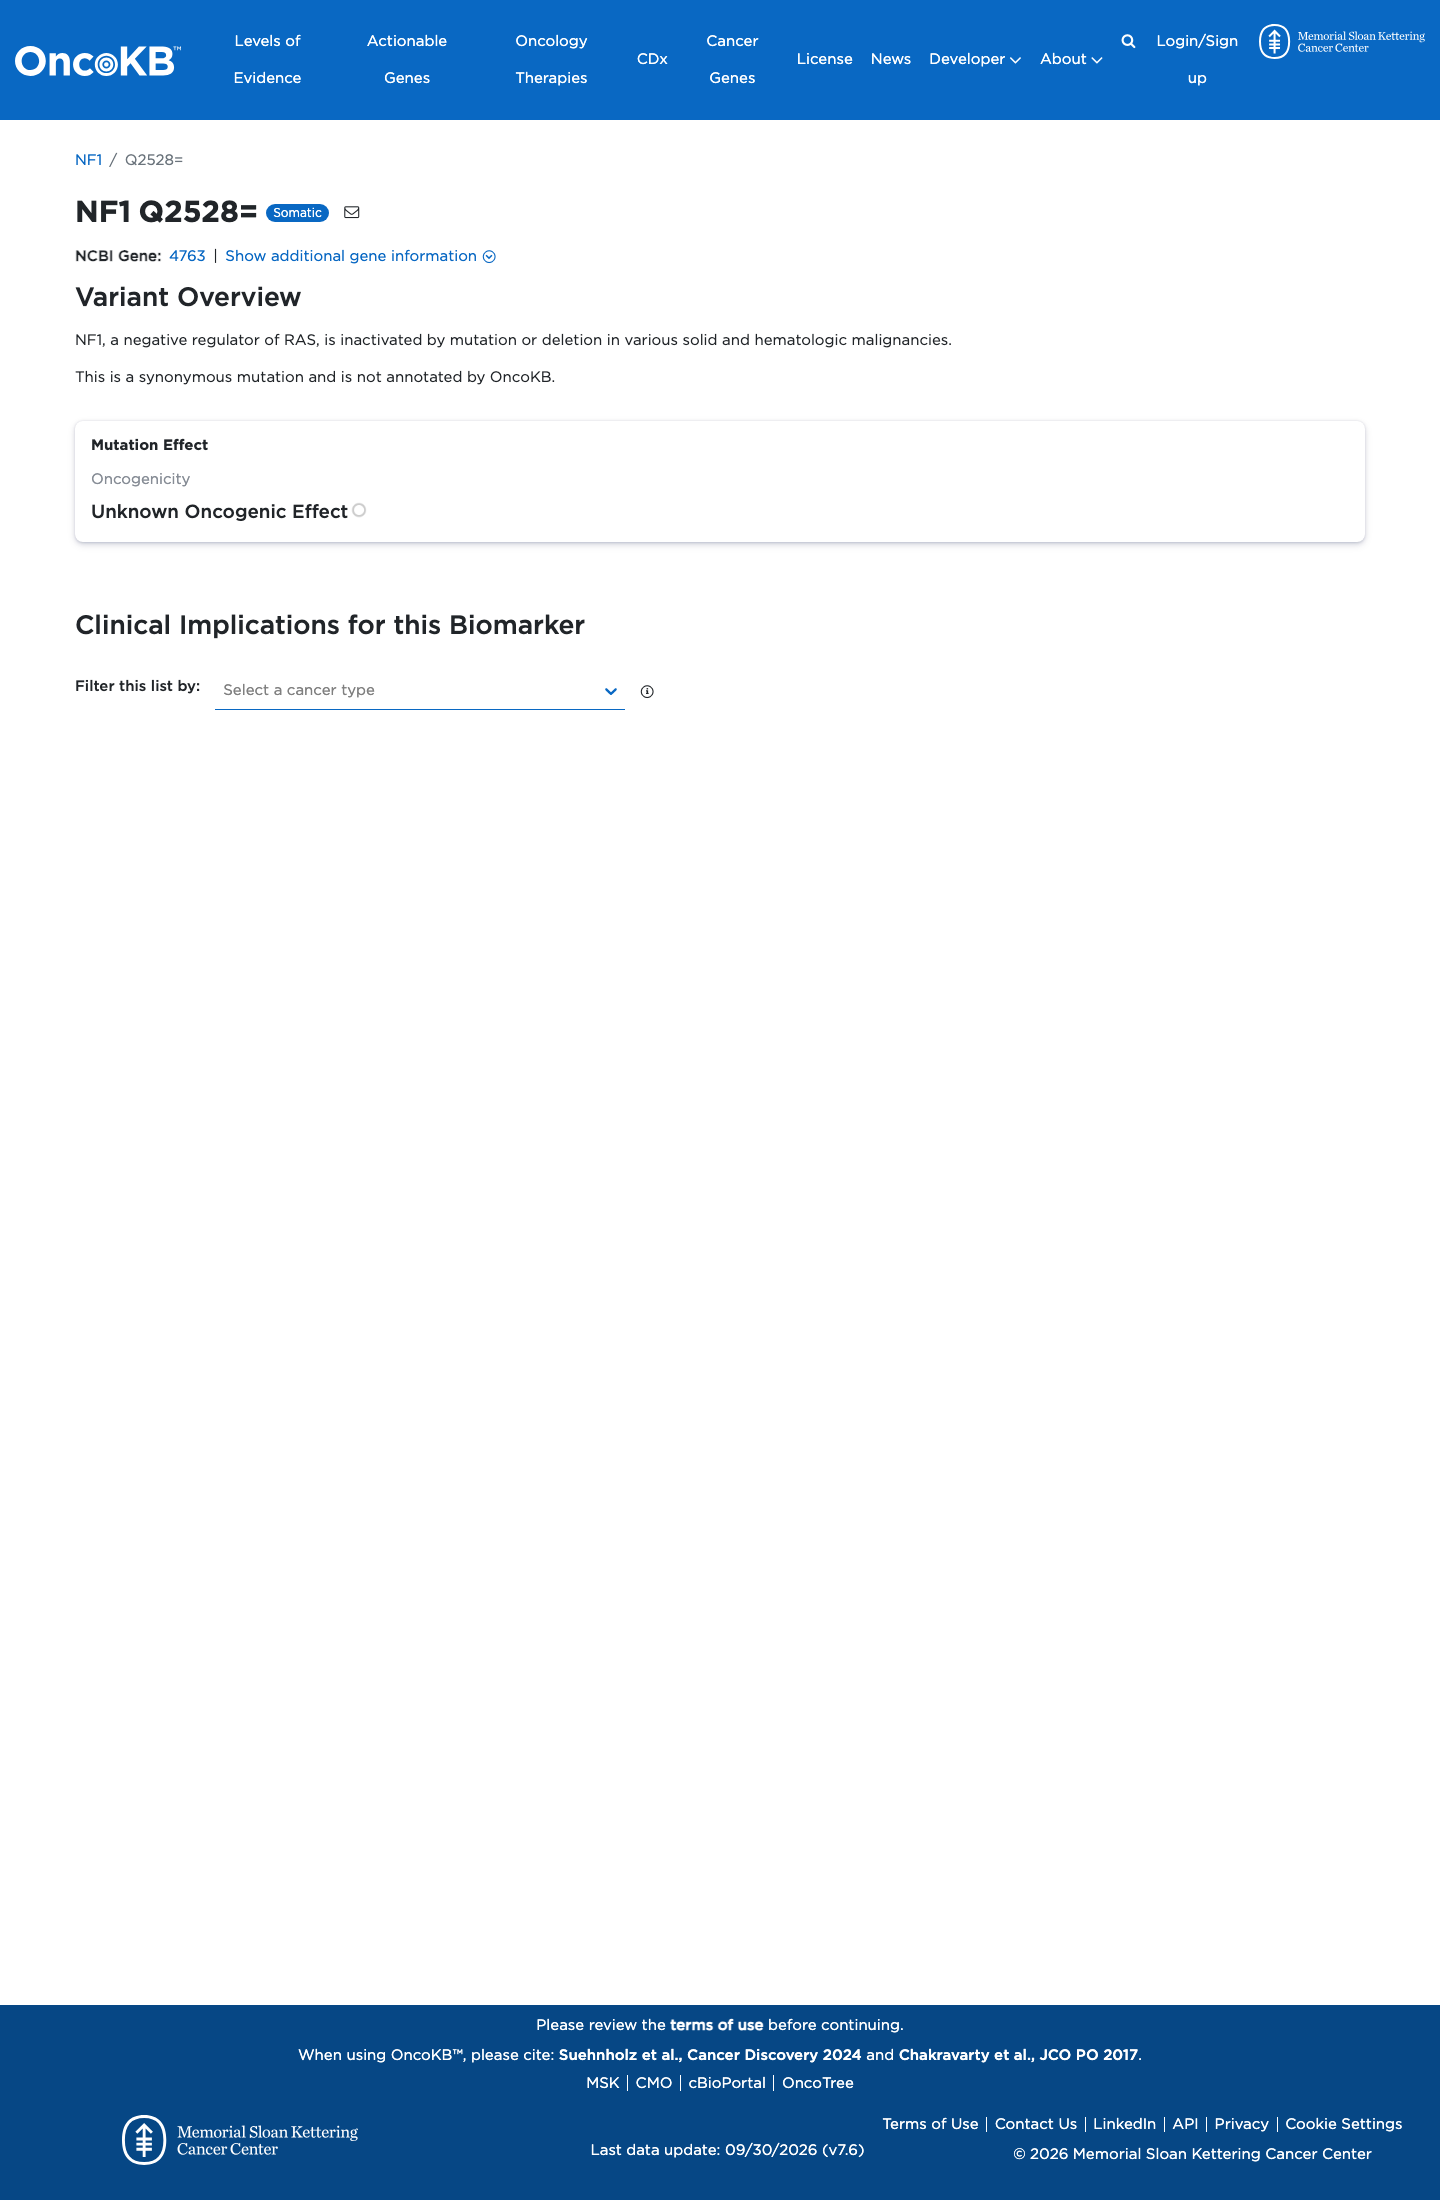
Functional / OncoKB screenshot
Functional Unknown Oncogenic Effect
OncoKB identified curated literature and non-variant-specific oncogenicity context for review; listed oncogenicity label: Unknown Oncogenic Effect.
OncoKB ↗
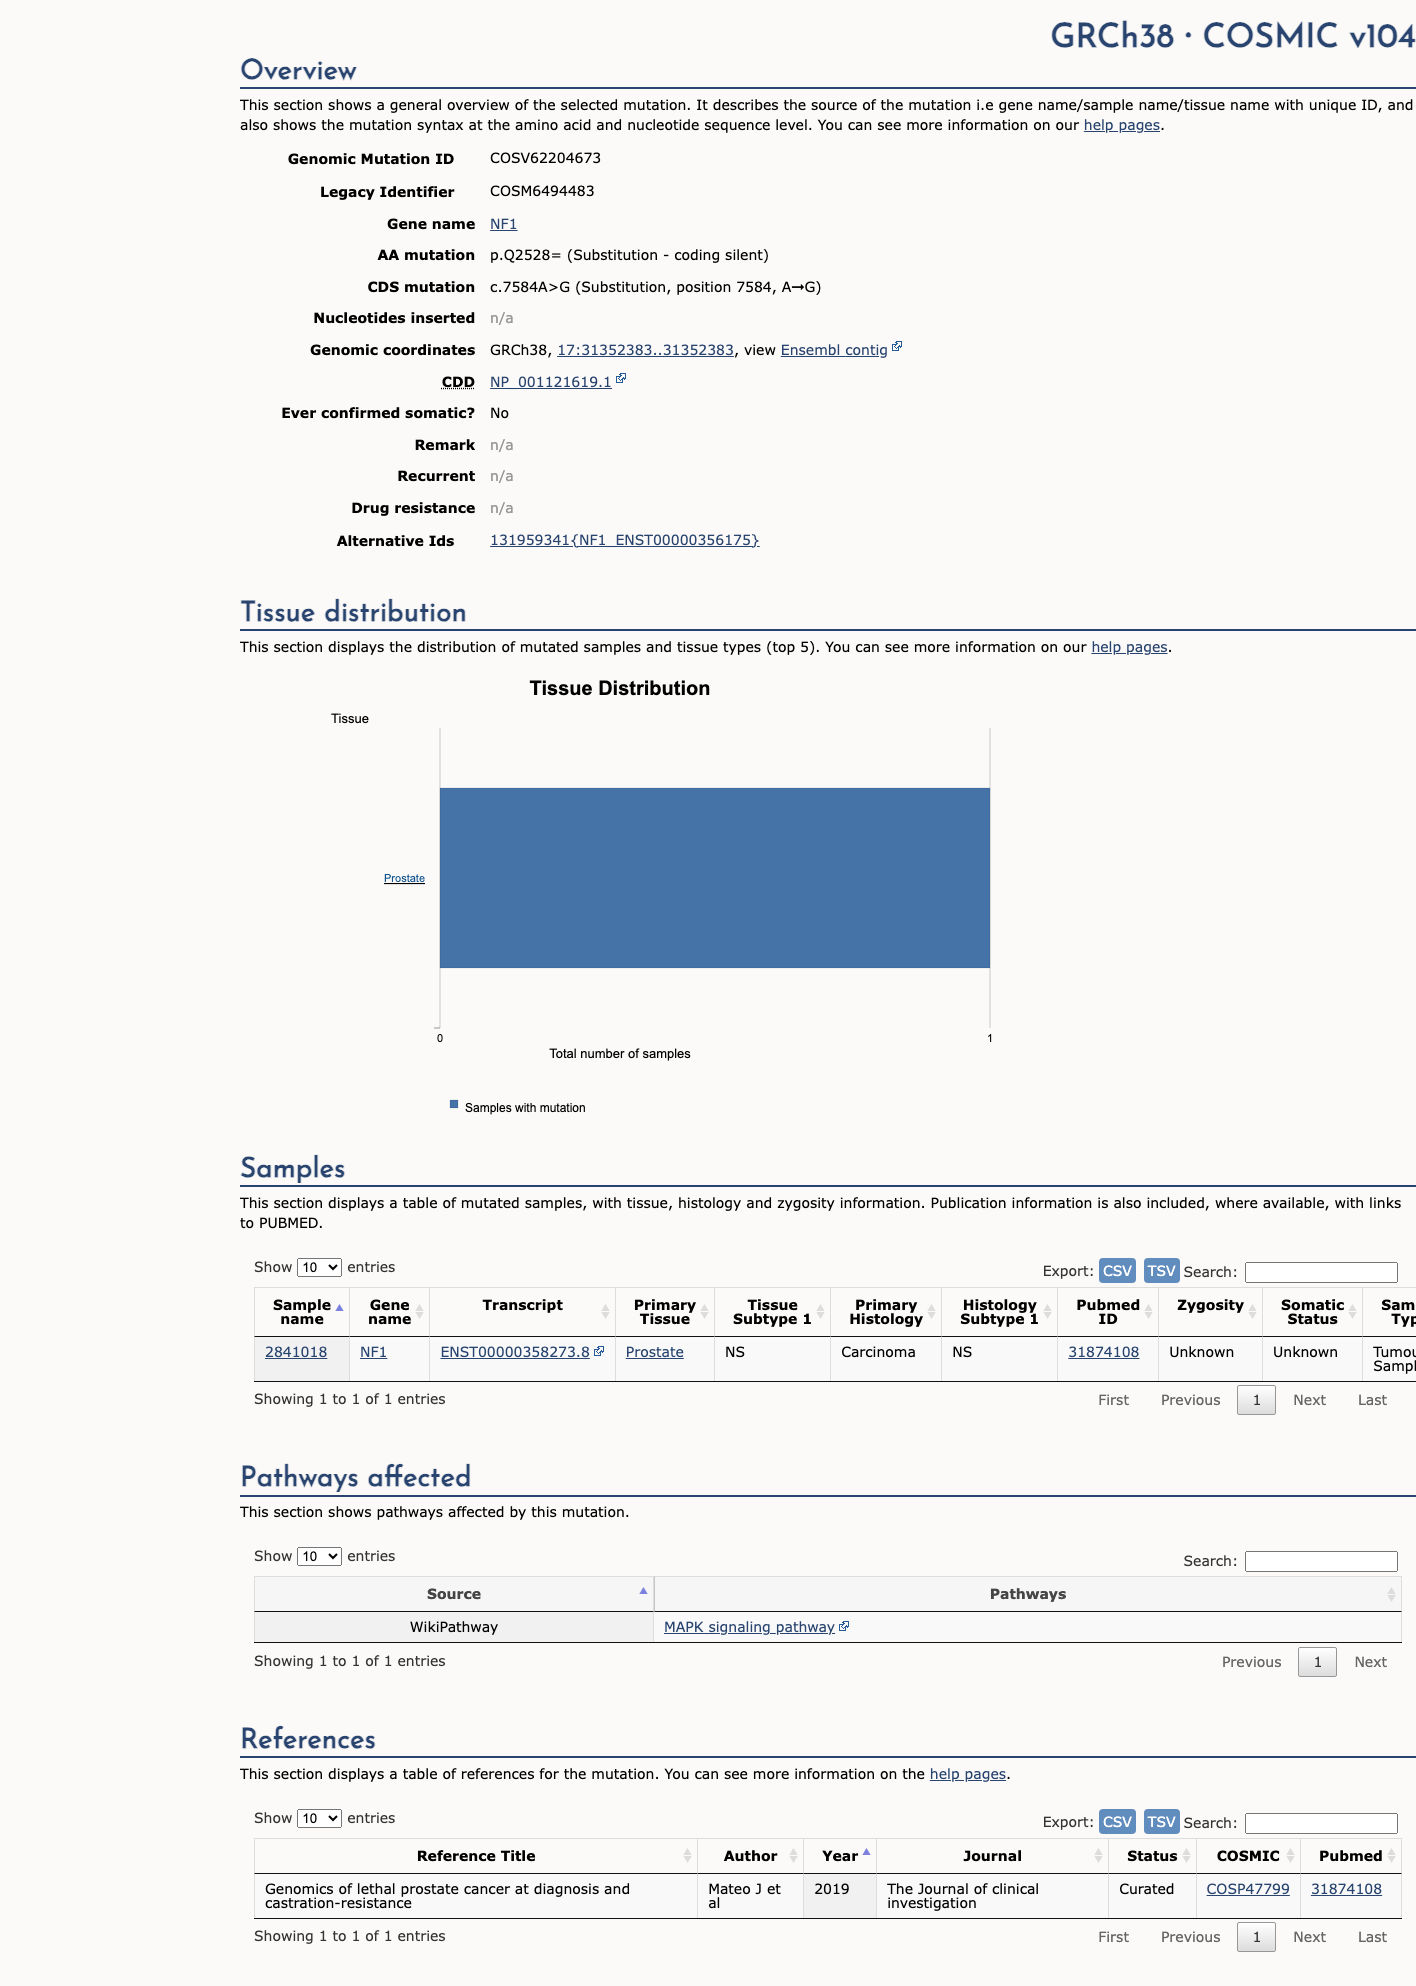
COSMIC screenshot
COSMIC
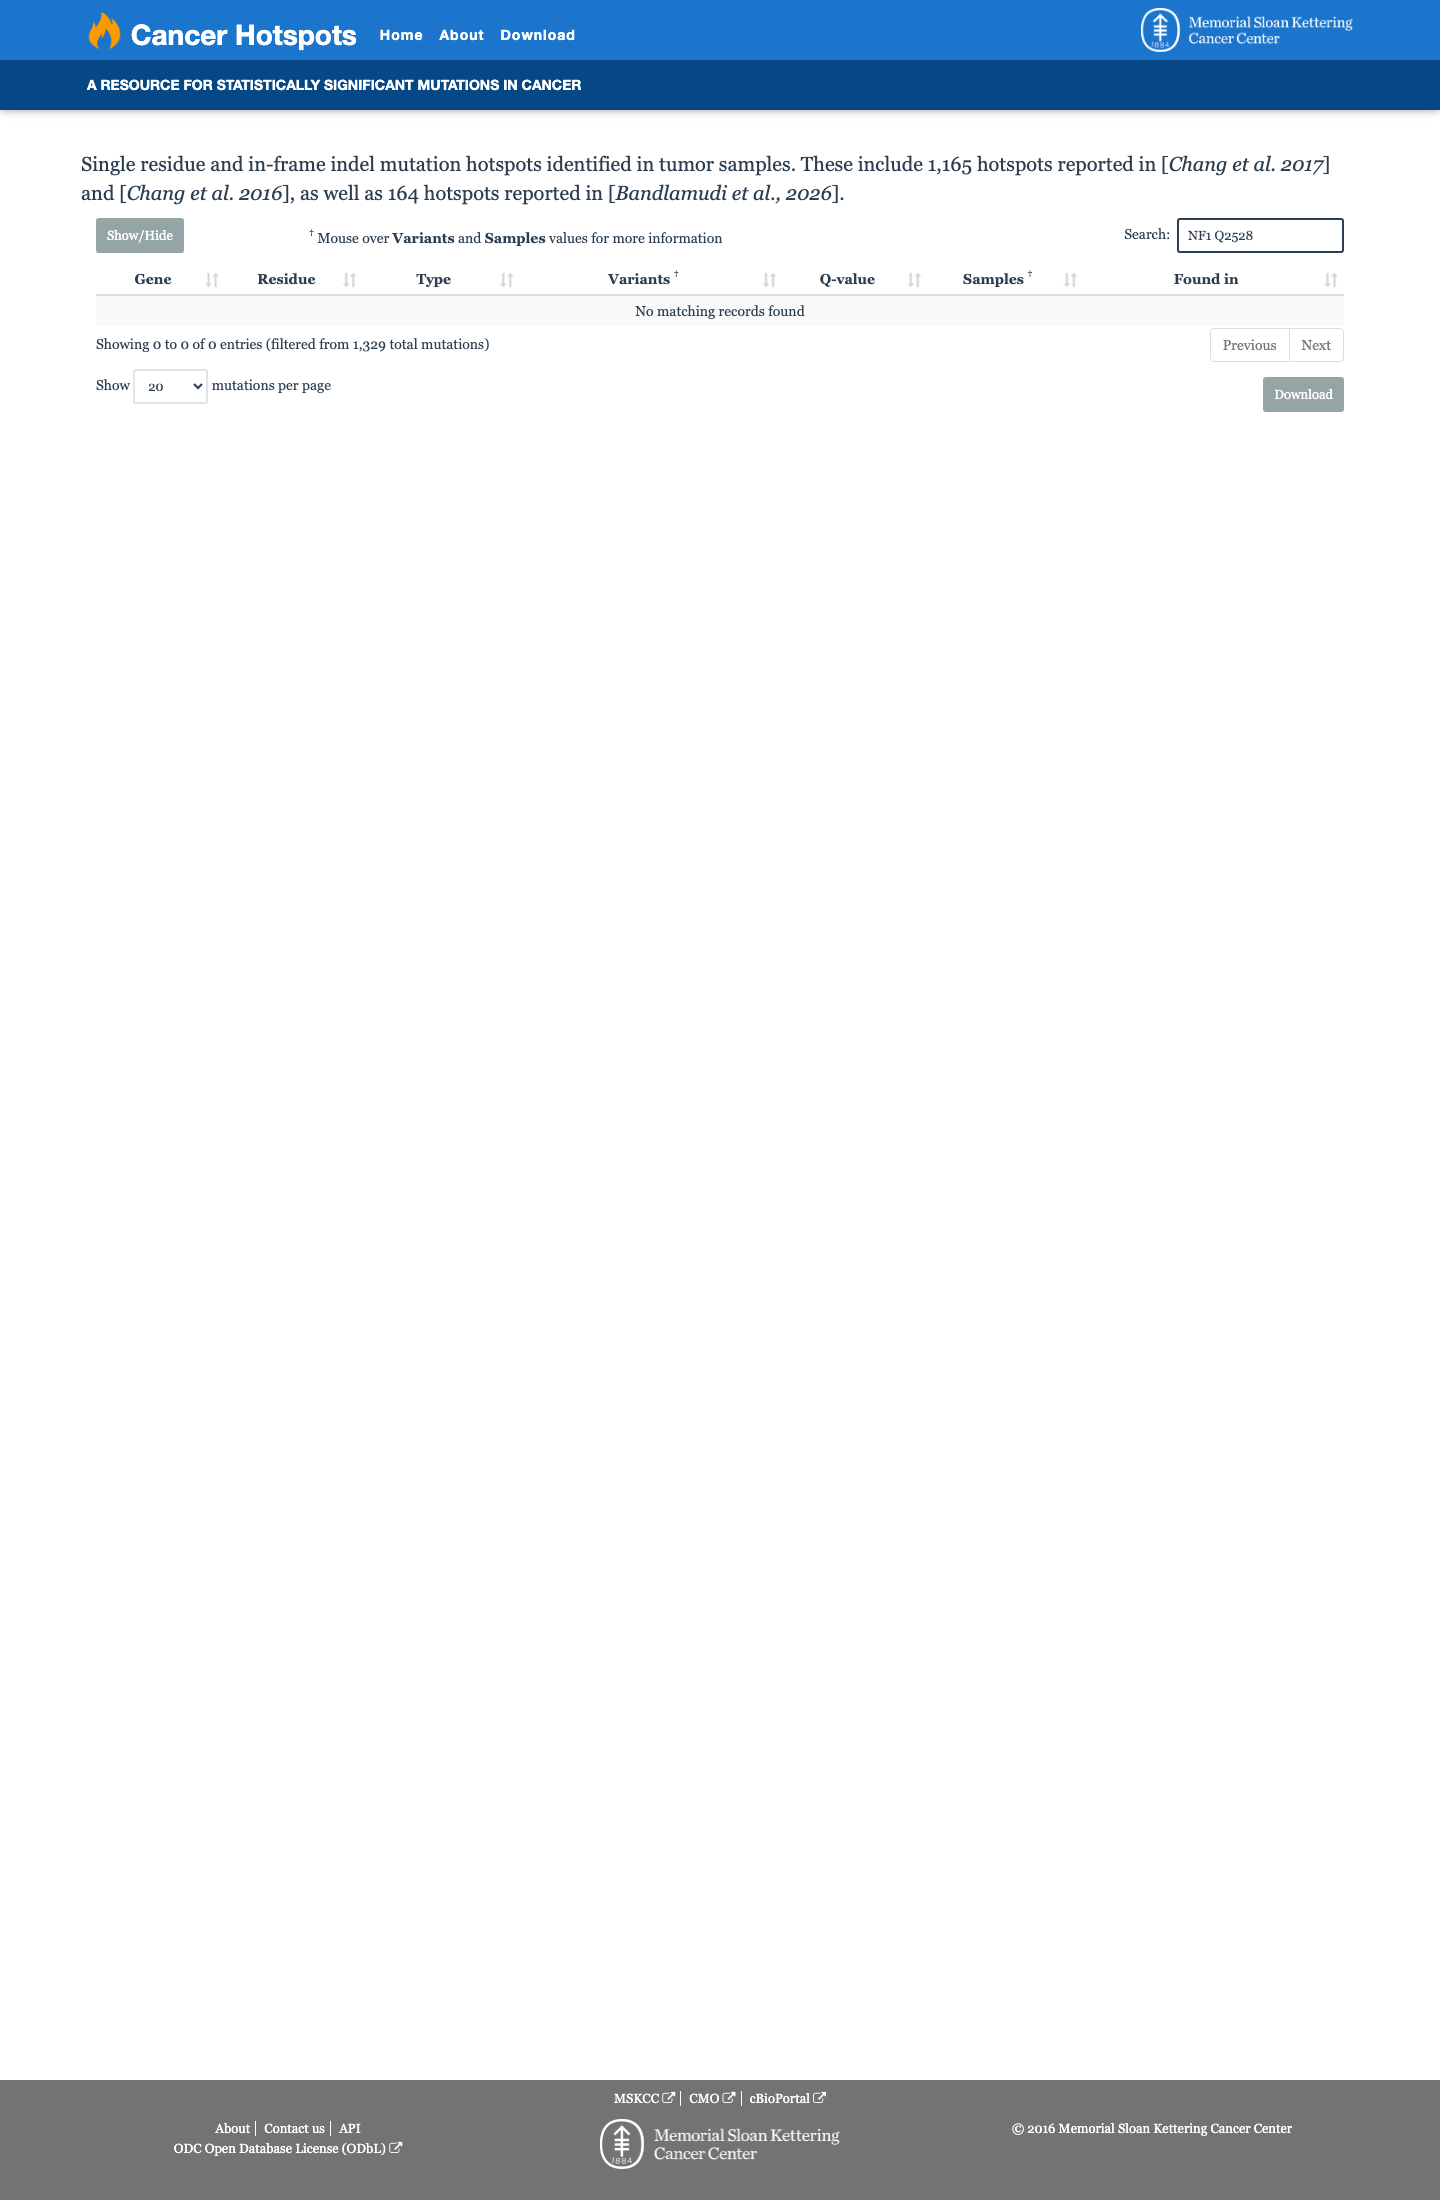
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has previously been reported in somatic cancers (COSMIC; COSV62204673, n = 1 times).
Hotspots
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
1papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 9 further PMIDs triaged but not cited — see Sources & references.
Rule & framework references · cited for criterion definitions, not variant evidence
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.
Sources & reference links
9Sources
CSpec VCEP
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 9 PMIDs not cited in assessment
23460398 ↗ Neurofibromatosis-1 gene deletions and mutations in de novo adult acute myeloid leukemia. CLINVAR
24033266 ↗ A systematic approach to assessing the clinical significance of genetic variants. CLINVAR
10678181 ↗ Nf1 and Gmcsf interact in myeloid leukemogenesis. CLINVAR
20065170 ↗ American Society of Clinical Oncology policy statement update: genetic and genomic testing for cancer susceptibility. CLINVAR
20301288 ↗ Neurofibromatosis 1. CLINVAR
26324357 ↗ American Society of Clinical Oncology Policy Statement Update: Genetic and Genomic Testing for Cancer Susceptibility. CLINVAR
26140447 ↗ Points to Consider: Ethical, Legal, and Psychosocial Implications of Genetic Testing in Children and Adolescents. CLINVAR
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification criteria. CLINVAR
33939658 ↗ The North American Neuroendocrine Tumor Society Consensus Guidelines for Surveillance and Management of Metastatic and/or Unresectable Pheochromocytoma and Paraganglioma. CLINVAR