Back
NM_001127510.3:c.221-29G>C
p.? · APC
0%
complete
Final classification
Benign
BA1BS1BS2BP4BP7
APC
c.221-29G>C
p.?
unknown · exon 4i

APC is a tumor suppressor gene whose protein acts as a key brake on the Wnt signaling pathway by helping mark the growth-promoting protein beta-catenin for destruction; it also participates in cell migration, adhesion, and apoptosis. Inherited mutations in APC cause familial adenomatous polyposis (FAP), an autosomal dominant condition characterized by numerous colorectal polyps and a very high risk of colorectal cancer, with related forms including attenuated FAP and gastric adenocarcinoma and proximal polyposis of the stomach (GAPPS). Somatic APC mutations are found in roughly 50-80% of sporadic colorectal cancers, where they act as early tumor-initiating events, and are also observed in some breast, stomach, and prostate cancers. Loss of APC function leaves Wnt signaling abnormally active, driving uncontrolled cell growth, which makes APC a central tumor suppressor in colorectal cancer.

This variant

APC is a tumor suppressor whose loss of function drives autosomal dominant, near-fully-penetrant familial adenomatous polyposis and early colorectal cancer, so the disease-relevant APC alleles are truncating or splice-disrupting changes; c.221-29G>C sits in intron 3, distant from the canonical splice consensus and the coding sequence, and is a common allele in non-cancer adult populations.

Transcript
NM_001127510.3
HGVS · transcript:coding
NM_001127510.3:c.221-29G>C
GRCh38
chr5:112767160 G>C
GRCh37
chr5:112102857 G>C
Benign: BA1 (stand-alone benign) at non-cancer gnomAD v2.1.1 exome popmax filtering AF 0.477% meets the APC InSiGHT VCEP Rule26 stand-alone benign branch.
Classification rationale
BA1BS1BS2BP4BP7 Benign
APC c.221-29G>C unknown · exon 4i

BA1 stand-alone benign: non-cancer gnomAD v2.1.1 exome popmax filtering AF 0.4767% is ~4.8-fold above the APC VCEP's 0.1% stand-alone benign ceiling. BS1 strong: the same popmax filtering AF clears the VCEP's 0.001% strong-benign ceiling, but is subsumed by BA1 and not added to the tally. BS2 strong: 3 homozygotes in gnomAD v2.1 non-cancer exomes (9 in v4.1) exceed the VCEP's >= 2 homozygote requirement for this near-fully-penetrant dominant disorder. BP4 supporting: SpliceAI max delta 0.02 and Pangolin near zero predict no impact for this intronic variant, below the 0.1 benign threshold. BP7 supporting: at -29, beyond the +7/-21 boundary, with multiple algorithms predicting no splice-consensus impact and no new splice site.

BA1 + BS1 + BS2 + BP4 + BP7 Benign
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_001127510.3 · variants mapped to exon structure
APC NM_001127510.3
Fetching transcript structure from UCSC…
Applied criteria · 5 applied · 14 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 5
Strength Supporting Moderate Strong Very strong
BA1 stand-alone Benign
Met: non-cancer gnomAD v2.1 exome popmax filtering AF 0.477% exceeds the VCEP BA1 threshold of 0.1%, giving stand-alone benign evidence.
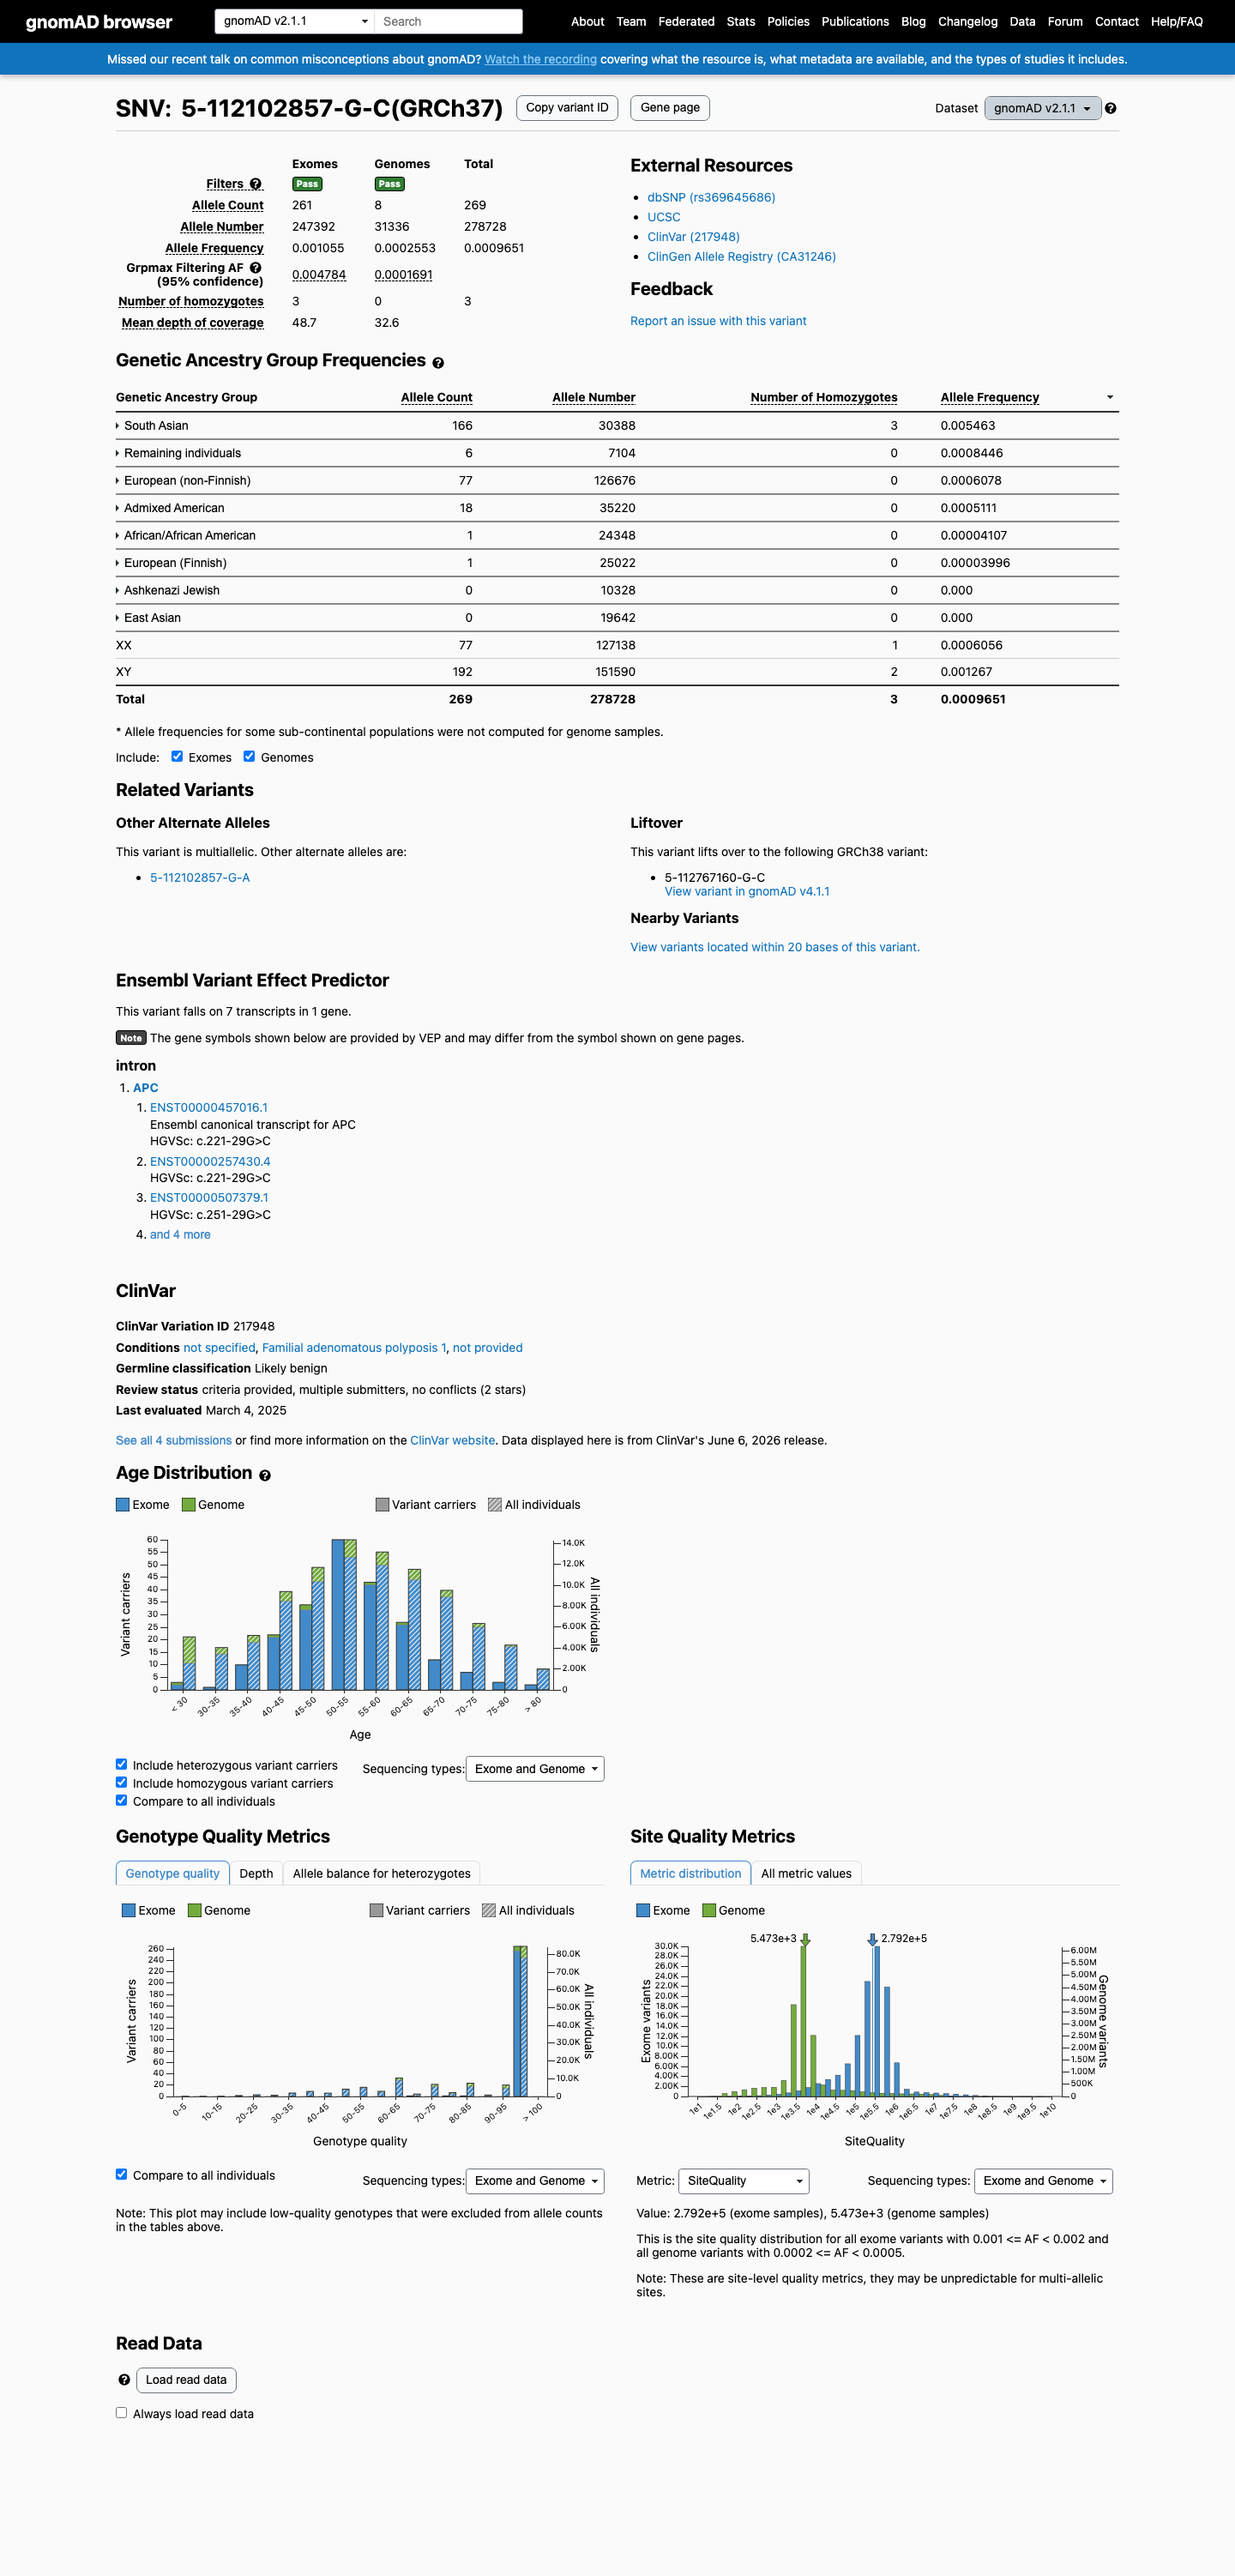
Governing specification: ClinGen InSiGHT Hereditary Colorectal Cancer/Polyposis Expert Panel Specifications for APC, version 2.1 (cspec). BA1 rule text: 'GnomAD Popmax Filtering Allele Frequency (AF) >= 0.1% (0.001)'; default strength 'Benign Stand Alone'; instructionsToUse: 'General recommendation: Use the non-cancer dataset from gnomAD (v2.1.1)'.gnomAD v2.1 non-cancer subset, exomes (same variant record page as registry key gnomad_v2, build GRCh37, variant id 5-112102857-G-C): 252/233,222 alleles, AF 0.00108052 (0.10805%), homozygotes = 3; South Asian 165/30,298 alleles, AF 0.00544590 (0.54459%), homozygotes = 3; non-cancer exome popmax (grpmax) filtering AF 0.00476693 (0.4767%). This popmax filtering AF exceeds the 0.1% BA1 cutoff.gnomAD v2.1 all-comers (gnomad_v2) for comparison: total AF 0.00096510 (0.09651%, 269/278,728 alleles, 3 homozygotes), exome AF 0.00105501 (0.10550%, 261/247,392 alleles), popmax filtering AF 0.00478413 (0.4784%), South Asian AF 0.00546268 (0.54627%).
BS1 strong Benign
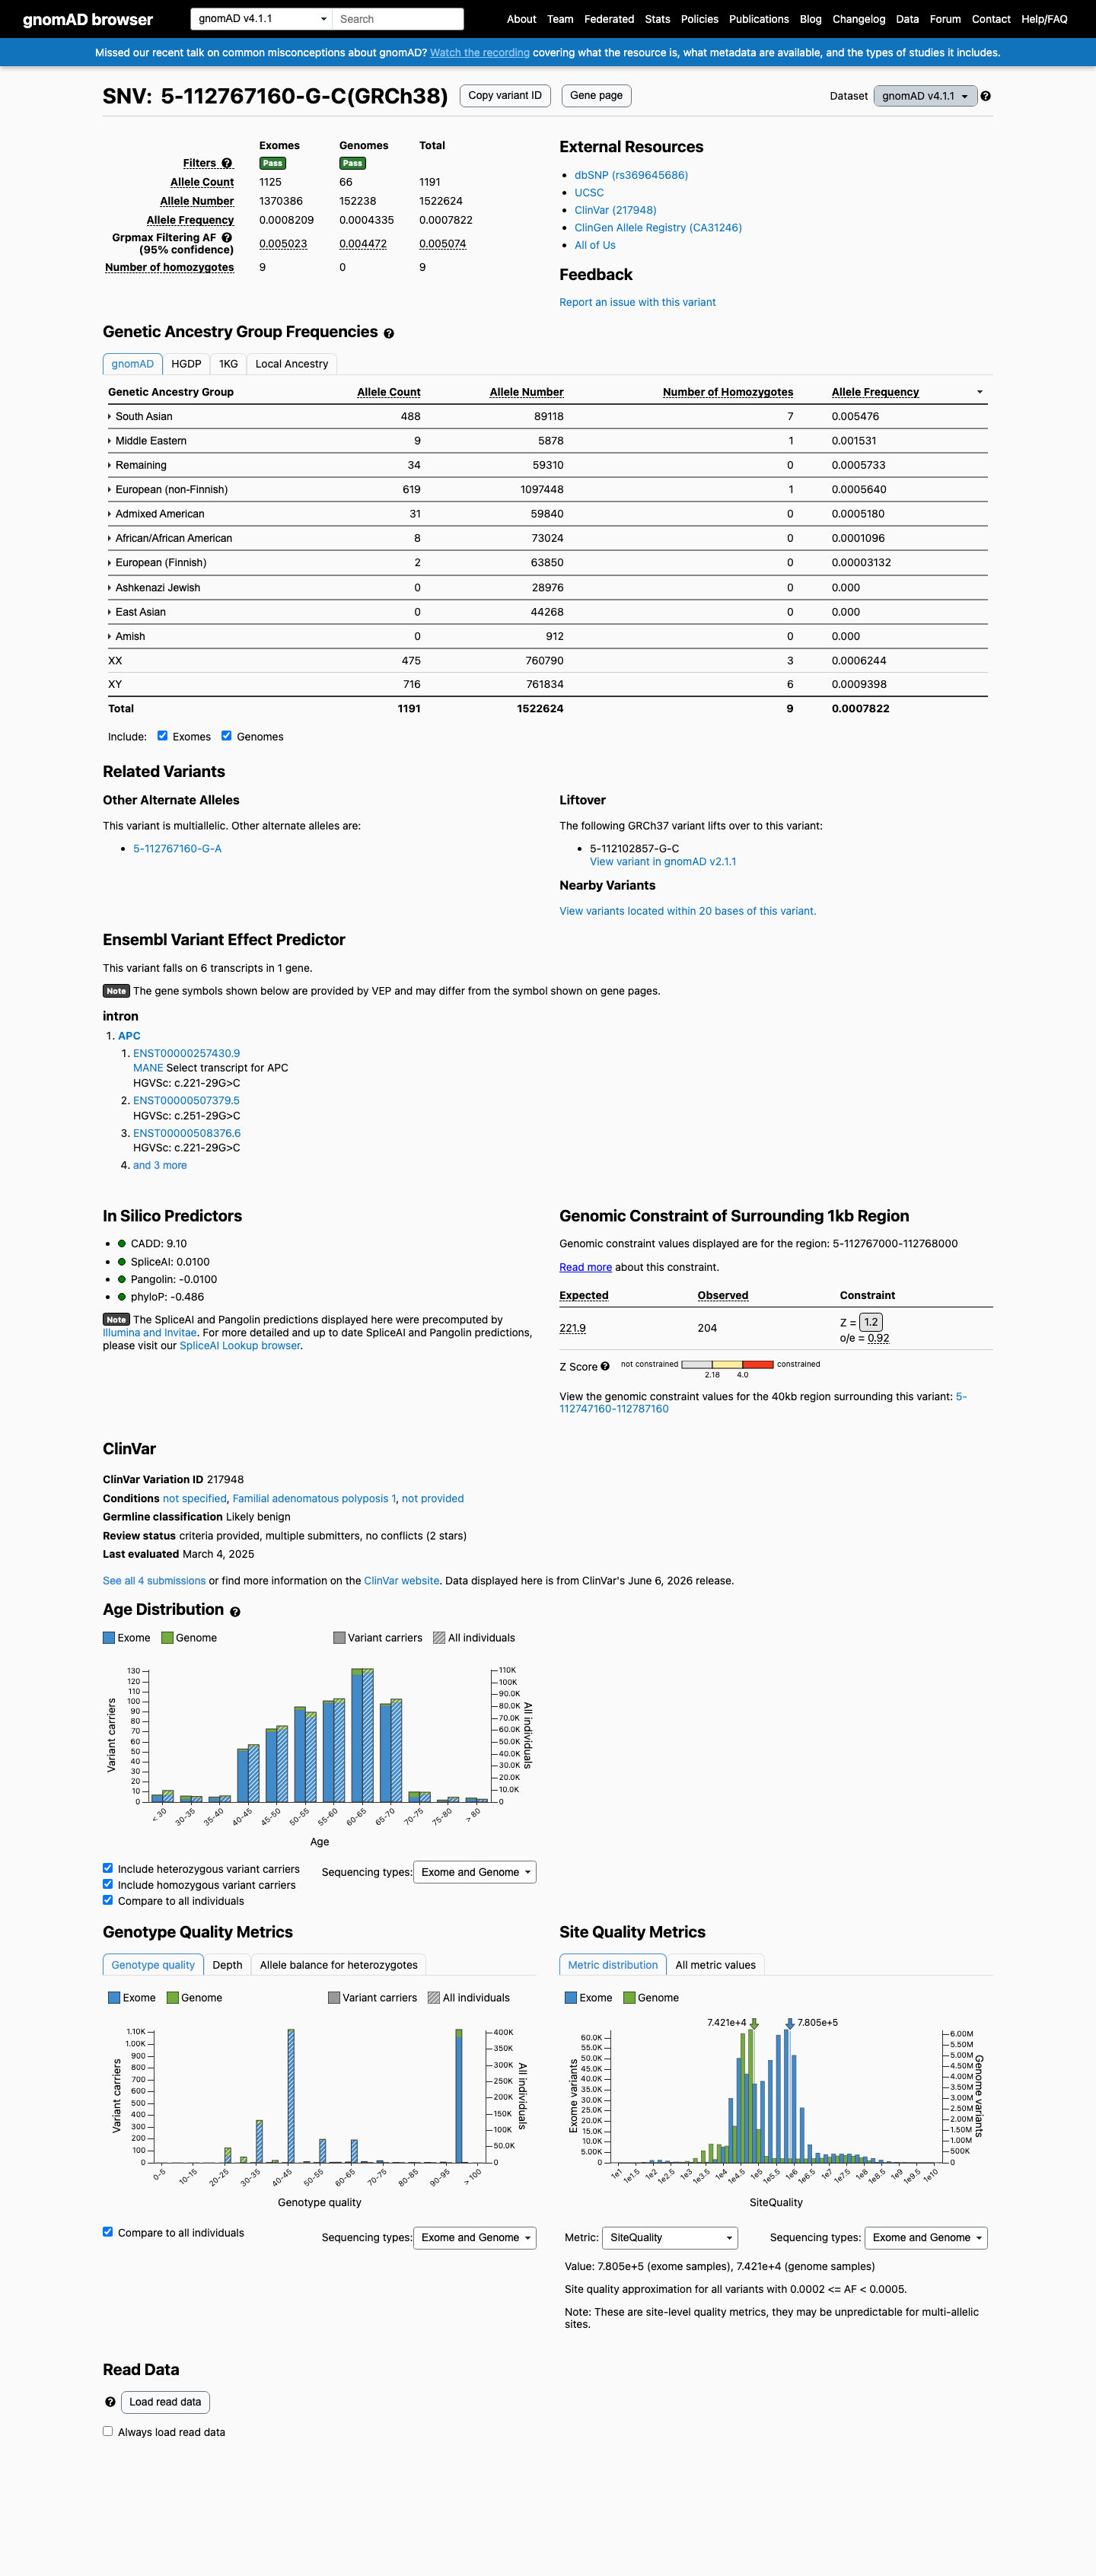
Met: the same non-cancer gnomAD v2.1 exome popmax filtering AF of 0.477% exceeds the VCEP BS1 threshold of 0.001%, but it is subsumed by BA1.
Governing specification: ClinGen InSiGHT APC specification v2.1 (cspec). BS1 rule text: 'GnomAD Popmax Filtering Allele Frequency (AF) >= 0.001% (0.00001)'; default strength 'Benign Strong'; instructionsToUse: 'General recommendation: Use the non-cancer dataset from gnomAD (v2.1.1)'.gnomAD v2.1 non-cancer exomes (gnomad_v2 record page): popmax (grpmax) filtering AF 0.00476693 (0.4767%) versus the 0.001% BS1 cutoff; allele frequency 0.10805% (252/233,222 alleles).gnomAD v2.1 all-comers popmax filtering AF 0.00478413 (0.4784%), total AF 0.09651% (269/278,728 alleles); gnomAD v4.1 popmax filtering AF 0.00507372 (0.5074%), total AF 0.07822% (1,191/1,522,624 alleles); gnomAD-Canada v1.0 AF 0.10859% (20/18,418 alleles). All exceed the 0.001% threshold.
BS2 strong Benign
Met: 3 homozygotes in gnomAD v2.1 non-cancer exomes exceed the VCEP BS2 requirement of at least 2 homozygous observations for this near-fully-penetrant dominant disorder.
Governing specification: ClinGen InSiGHT APC specification v2.1 (cspec). BS2 strong rule text: '>= 10 points for healthy individuals OR >= 2 times in homozygous state'; instructionsToUse: 'The non-cancer dataset from gnomAD (v2.1.1) cannot be used for heterozygous healthy individuals, because of the limited phenotype information and since it is usually already used for BA1/BS1. However, the non-cancer dataset from gnomAD (v2.1.1) can be used to search for homozygous individuals.'gnomAD v2.1 non-cancer exomes (gnomad_v2 record page): homozygote count = 3 (233,222 alleles; 252 allele carriers), with all 3 homozygotes in the South Asian subpopulation (165/30,298 alleles, homozygote count 3). Three homozygous observations satisfy the specification's >= 2 requirement.gnomAD v4.1 (gnomad_v4) independent corroboration: 9 homozygotes in total (7 South Asian, 1 Middle Eastern, 1 non-Finnish European) among 1,522,624 alleles.
BP4 supporting Benign
Met at supporting: intronic c.221-29 variant with SpliceAI max delta 0.02 and Pangolin near zero, both below the 0.1 BP4 threshold.
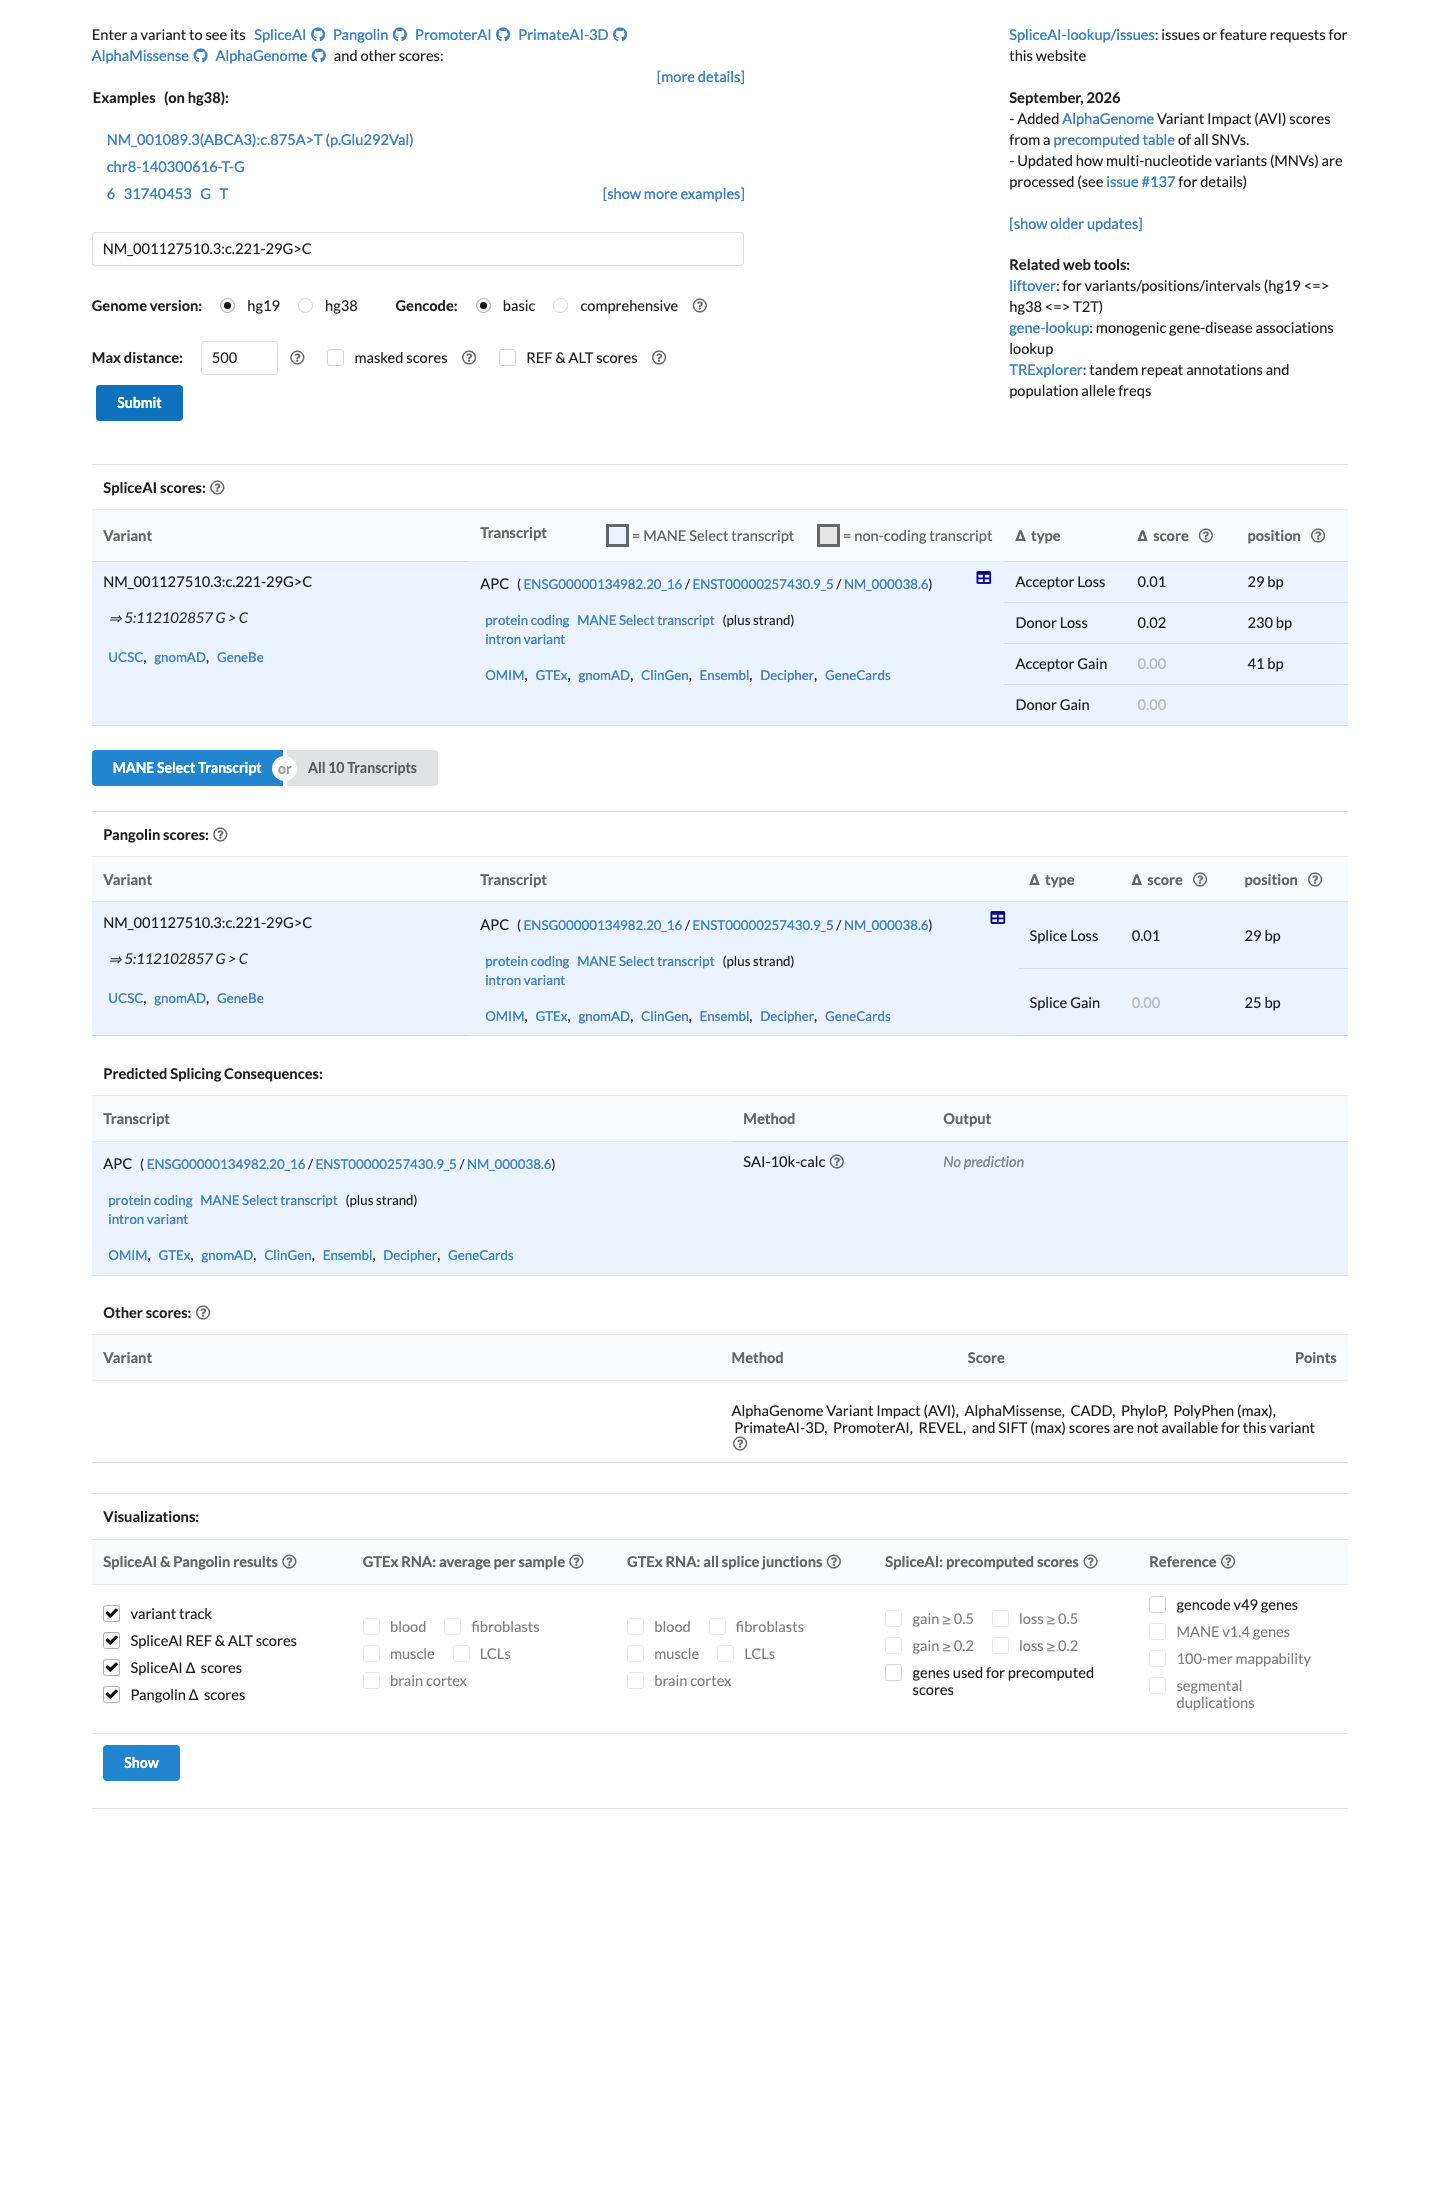
APC VCEP v2.1 BP4 (benign supporting) rule: 'Missense variants: BP4 is not applicable. Synonymous (silent) or intronic variants: Multiple in silico splicing predictors suggest no impact on gene or gene product.' Recommended programs are referenced to the panel's PS1 instructions (SpliceAI, MaxEntScan, VarSeak).Variant NM_001127510.3:c.221-29G>C is intronic (29 nucleotides upstream of the exon 3 acceptor; NC_000005.10:g.112767160G>C), which places it in BP4's synonymous/intronic scope rather than the inapplicable missense scope.SpliceAI Lookup result: max delta 0.02 (DS_AL 0.013, DS_DL 0.020, DS_AG 0.001, DS_DG 0.000) with Pangolin SG 0.001 and Pangolin SL -0.007 - two independent algorithms agree that the native splice sites are unaffected and no cryptic site is created.
BP7 supporting Benign
Met at supporting: intronic at -29 (beyond the -21 boundary) with SpliceAI max delta 0.02 and Pangolin near zero, showing no splice-site impact.
APC VCEP v2.1 BP7 (benign supporting) rule: 'A synonymous (silent) or intronic variant at or beyond +7/-21 for which multiple splicing prediction algorithms predict no impact to the splice consensus sequence nor the creation of a new splice site.' This gene-specific wording extends BP7 beyond the generic synonymous-only definition and therefore covers the intronic variant assessed here.NM_001127510.3:c.221-29G>C sits 29 nucleotides upstream of the exon 3 acceptor site (NC_000005.10:g.112767160G>C), i.e. beyond the -21 cutoff, so its position is inside the VCEP's BP7 scope; it is not a canonical +/-1 or 2 splice site (the APC VCEP's PVS1 decision-tree lists cover only those positions).The APC VCEP supplementary material records that the APC BP7 rule was modified from the ClinGen CDH1 specification and that use of BP7 together with BP4 is allowed.
Assessed · not applied · 5 not met · 9 not assessed
Pathogenic
PS1 Not met: no previously established pathogenic variant shares this nucleotide, and SpliceAI max delta 0.02 is far below splice-supportive thresholds.
PS2 Not assessed: no proband-level parental testing or maternity/paternity confirmation exists to compute an APC de novo score.
PS3 Not assessed: no RNA, minigene, or protein assay result exists for c.221-29G>C, and the APC VCEP requires RNA assay data showing a premature stop or exon 13/14 skipping.
PS4 Not assessed: no affected carrier with any Table 1 phenotype feature is reported, so the VCEP >=1 phenotype point PS4 minimum cannot be scored.
PM2 Not met: non-cancer exome allele frequency 0.108% (252 of 233,222 alleles) far exceeds the VCEP PM2 ceiling of 0.0003% for alleles counted more than once.
PM6 Not assessed: no assumed-de-novo proband observation or parental testing data exists to compute an APC de novo score.
PP1 Not assessed: no pedigrees, affected relatives, or meiosis counts are reported to meet the >=3-meiosis APC PP1 threshold.
PP3 Not met: SpliceAI max delta 0.02 and Pangolin near zero, both below the 0.2 supporting threshold for a deleterious splice effect.
PP5 Not met: ClinVar 217948 has zero expert-panel submissions and only 2-star laboratory assertions, so the expert-panel requirement for PP5 is not satisfied.
Benign
BS3 Not assessed: no RNA or beta-catenin reporter assay exists for c.221-29G>C, so the APC VCEP BS3 requirement for an assay showing no mRNA aberration is unmet.
BS4 Not assessed: no affected noncarrier with an APC phenotype score is reported to meet the BS4 threshold.
BP2 Not assessed: no source reports c.221-29G>C in trans with a pathogenic APC variant, and no phase data exist to satisfy the VCEP BP2 rule.
BP5 Not assessed: no colorectal polyposis phenotype and no variant in another polyposis gene is documented, so the VCEP BP5 condition cannot be evaluated.
BP6 Not met: the ClinVar Likely benign label rests on ordinary laboratory submissions with zero expert-panel assertions, which cannot trigger BP6.
N/A · 9 PVS1 · PM1 · PM3 · PM4 · PM5 · PP2 · PP4 · BP1 · BP3
Research & evidence
Population frequency · supports benign
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 0.000782202; MAF= 0.07822%, 1191/1522624 alleles, homozygotes = 9) and has highest observed frequency in the South Asian population (AF= 0.00547589; MAF= 0.54759%, 488/89118 alleles, homozygotes = 7); grpmax FAF= 0.00507372.
v2.1
This variant is present in gnomAD v2.1 (AF= 0.000965099; MAF= 0.09651%, 269/278728 alleles, homozygotes = 3) and has highest observed frequency in the South Asian population (AF= 0.00546268; MAF= 0.54627%, 166/30388 alleles, homozygotes = 3); grpmax FAF= 0.00478413.
🇨🇦 CA
This variant is present in gnomAD-Canada v1.0 (AF= 0.001085894233901618, 20/18418 alleles, homozygotes = 0).
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.078% · 1191 / 1,522,624
9 hom · FAF 0.51%
South Asian
488 / 89,118
0.55%
7 hom
Middle Eastern
9 / 5,878
0.15%
1 hom
Remaining individuals
34 / 59,310
0.057%
European (non-Finnish)
619 / 1,097,448
0.056%
1 hom
Admixed American
31 / 59,840
0.052%
African/African American
8 / 73,024
0.011%
European (Finnish)
2 / 63,850
0.0031%
+ 3 not observed (Amish, East Asian, Ashkenazi Jewish)
gnomAD v2.1
0.097% · 269 / 278,728
3 hom · FAF 0.48%
South Asian
166 / 30,388
0.55%
3 hom
Remaining individuals
6 / 7,104
0.084%
European (non-Finnish)
77 / 126,676
0.061%
Admixed American
18 / 35,220
0.051%
African/African American
1 / 24,348
0.0041%
European (Finnish)
1 / 25,022
0.004%
+ 2 not observed (Ashkenazi Jewish, East Asian)
gnomAD Canada 🇨🇦
0.11% · 20 / 18,418
0 hom · FAF 0.19%
South Asian
6 / 1,362
0.44%
Remaining individuals
3 / 1,138
0.26%
Latino/Admixed American
2 / 838
0.24%
European (non-Finnish)
9 / 11,738
0.077%
+ 5 not observed (African/African American, Ashkenazi Jewish, East Asian, European (Finnish), Middle Eastern)
ClinVar screenshot
ClinVar
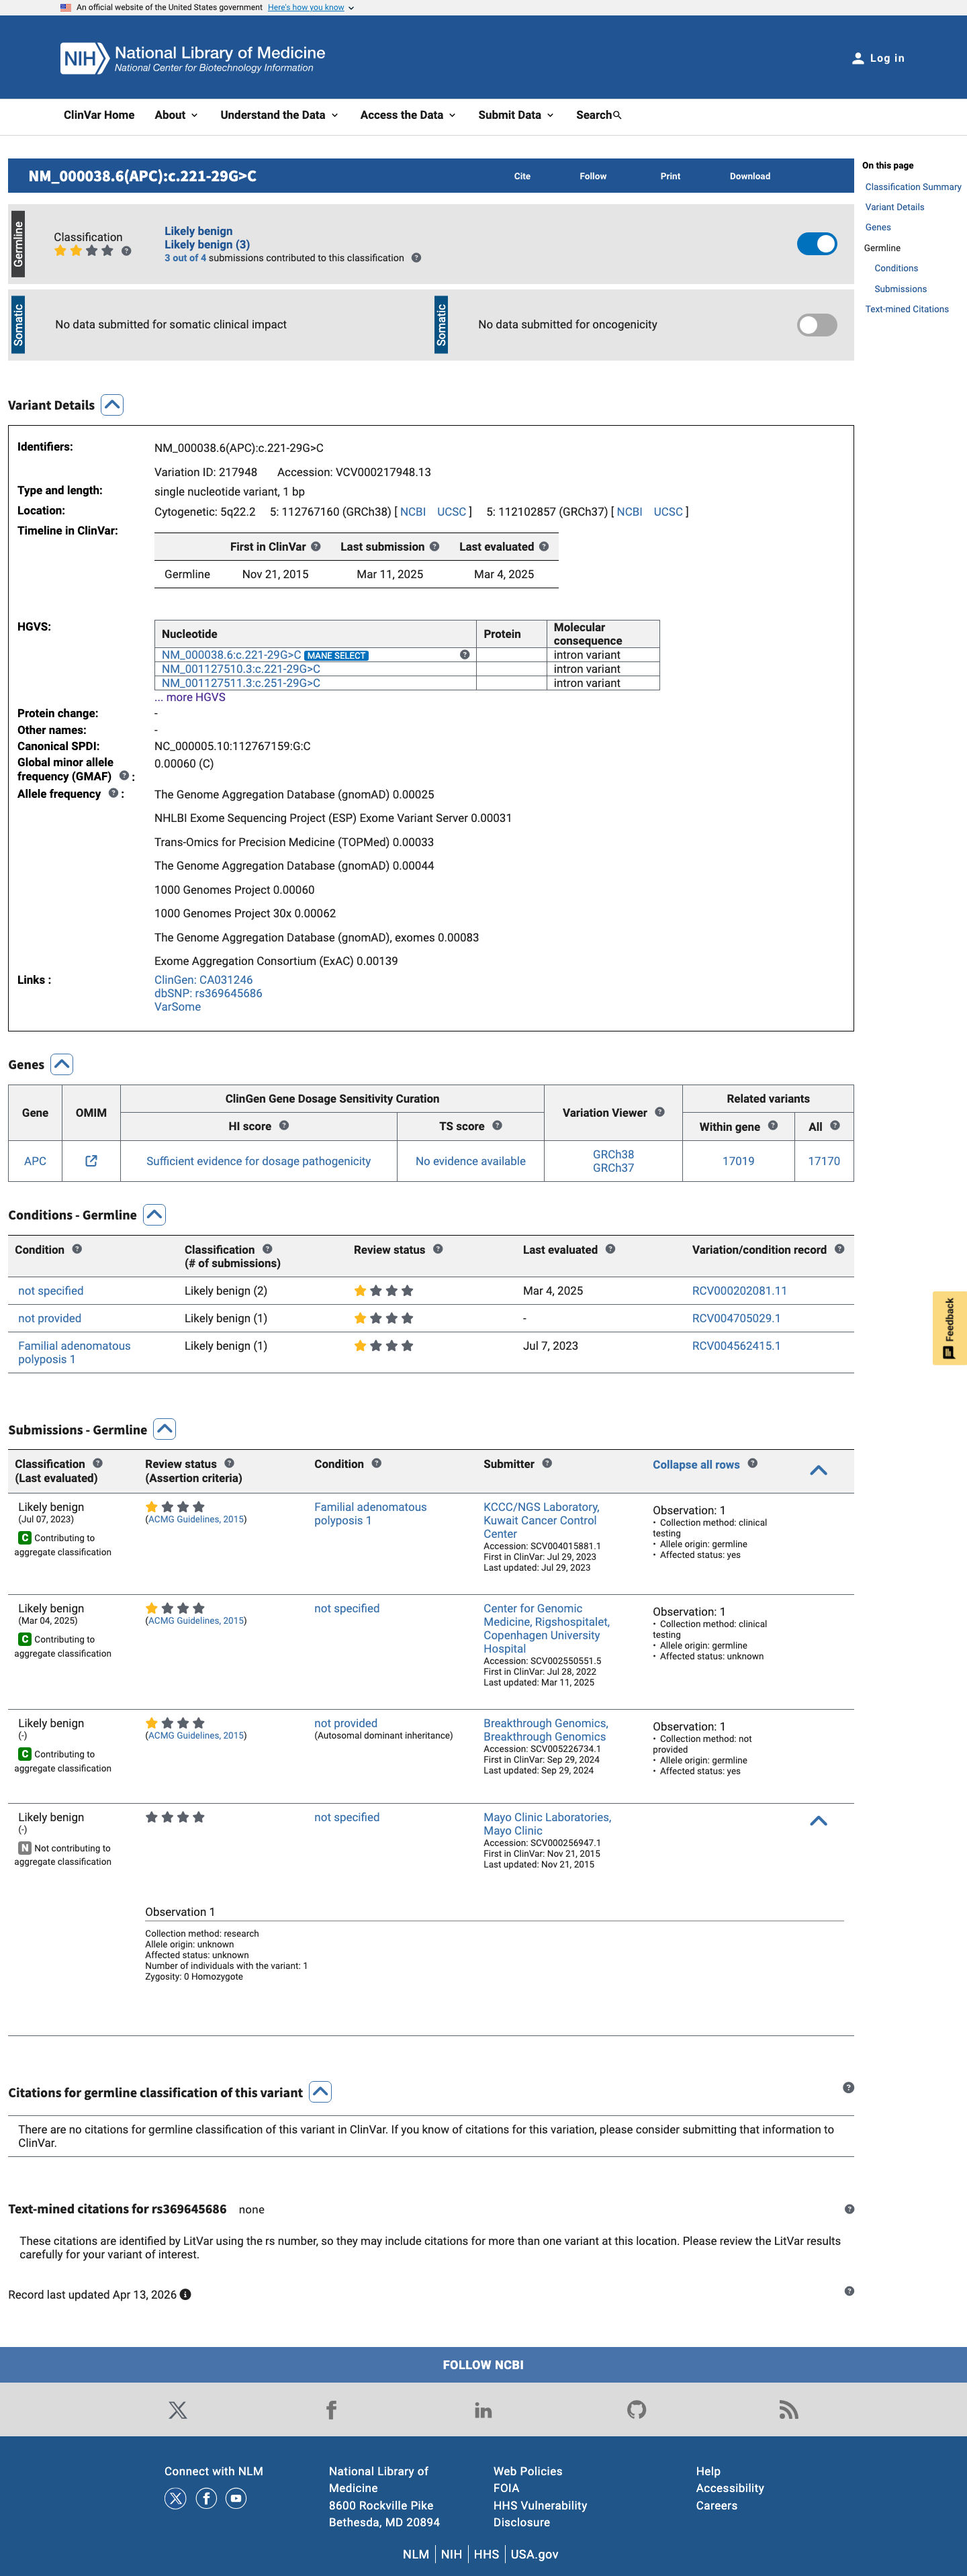
This variant has been reported in ClinVar as Likely benign (2 clinical laboratories). (ClinVarID = 217948)
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.02).
Functional No data
No calibrated functional assay or RNA evidence was identified for this variant.
OncoKB ↗
COSMIC screenshot
COSMIC
Somatic evidence
COSMIC
This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant cancer hotspot.
COSMIC ↗
Literature · how each cited paper was used
2papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 4 further PMIDs triaged but not cited — see Sources & references.
Rule & framework references · cited for criterion definitions, not variant evidence
25645574 ↗ ACG clinical guideline: Genetic testing and management of hereditary gastrointestinal cancer syndromes.
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.
Sources & reference links
8Sources
CSpec VCEP
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Triaged references · 4 PMIDs not cited in assessment
20301519 ↗ APC-Associated Polyposis Conditions. CLINVAR
21368914 ↗ Clinical utility gene card for: familial adenomatous polyposis (FAP) and attenuated FAP (AFAP). CLINVAR
26389258 ↗ Cancer Genetics Risk Assessment and Counseling (PDQ®): Health Professional Versi CLINVAR
27854360 ↗ Recommendations for reporting of secondary findings in clinical exome and genome sequencing, 2016 update (ACMG SF v2.0): a policy statement of the American College of Medical Genetics and Genomics. CLINVAR