Back
NM_001127511.2:c.78C>A
p.Ser26Arg · APC
0%
complete
Final classification
Benign
BA1BS1BS2BP1
APC
c.78C>A
p.Ser26Arg
missense · exon 1

APC is a tumor suppressor gene whose protein acts as a key brake on the Wnt signaling pathway by helping mark the growth-promoting protein beta-catenin for destruction; it also participates in cell migration, adhesion, and apoptosis. Inherited mutations in APC cause familial adenomatous polyposis (FAP), an autosomal dominant condition characterized by numerous colorectal polyps and a very high risk of colorectal cancer, with related forms including attenuated FAP and gastric adenocarcinoma and proximal polyposis of the stomach (GAPPS). Somatic APC mutations are found in roughly 50-80% of sporadic colorectal cancers, where they act as early tumor-initiating events, and are also observed in some breast, stomach, and prostate cancers. Loss of APC function leaves Wnt signaling abnormally active, driving uncontrolled cell growth, which makes APC a central tumor suppressor in colorectal cancer.

This variant

APC is a tumour suppressor whose loss of function leaves Wnt signalling abnormally active and causes autosomal dominant familial adenomatous polyposis and a high colorectal cancer risk; this common N-terminal missense change (p.Ser26Arg), seen at up to 2.9% allele frequency with multiple homozygotes in population datasets, is not the truncating loss-of-function allele type that drives APC-associated polyposis.

Transcript
NM_001127511.2
HGVS · transcript:coding
NM_001127511.2:c.78C>A
GRCh38
chr5:112707795 C>A
GRCh37
chr5:112043492 C>A
Benign: BA1 (stand-alone benign, non-cancer gnomAD popmax filtering AF 2.78%) alone satisfies the APC Expert Panel's Rule26.
Classification rationale
BA1BS1BS2BP1 Benign
APC c.78C>A missense · exon 1

BA1 stand-alone benign: non-cancer gnomAD v2.1.1 popmax filtering AF 2.78% is about 28-fold above the APC VCEP 0.1% threshold, which alone establishes Benign. BS1 strong: the same 2.78% popmax filtering AF is roughly 2,780-fold above the 0.001% threshold, though subsumed by BA1. BS2 strong: the APC VCEP homozygous-state clause is met, with 18 homozygotes in gnomAD v3.1 non-cancer genomes and 33 in v4.1 against a requirement of 2. BP1 supporting: a missense change at codon 26, outside the VCEP's excluded beta-catenin repeat (codons 1021-1035), in a gene where over 90% of pathogenic variants truncate.

BA1 + BS1 + BS2 + BP1 → Benign
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_001127511.2 · variants mapped to exon structure
APC NM_001127511.2
Fetching transcript structure from UCSC…
Applied criteria · 4 applied · 14 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 4
Strength Supporting Moderate Strong Very strong
BA1 stand-alone Benign
Met: non-cancer gnomAD v2.1.1 popmax filtering AF 2.78%, about 28-fold above the 0.1% APC VCEP stand-alone BA1 threshold.
APC VCEP specification v2.1 (InSiGHT) BA1 (Benign Stand Alone) rule: 'GnomAD Popmax Filtering Allele Frequency (AF) >= 0.1% (0.001)', with instructions to use the non-cancer dataset from gnomAD (v2.1.1); the supplementary material adds 'preferably based on the Popmax Filtering Allele Frequency of the non-cancer dataset from gnomAD (v2.1.1)' and permits subpopulations with >=2000 tested alleles, considering the highest minor allele frequency.Threshold derivation stated in the specification: prevalence 1:5,000, genetic heterogeneity 0.5, penetrance 0.8, allelic heterogeneity 0.06 (Kelly et al. 2018 plus the CardioDB allele frequency calculator) giving a calculated allele frequency of >=0.006%, rounded up 10-fold to >=0.1%.gnomAD v2.1 non-cancer exomes (VCEP-directed source, exome-only figures): popmax/exome grpmax filtering AF 0.02778749 (2.78%); AFR 0.03018054 (3.018%, 448/14844 alleles); allele frequency 0.00186391 (249/133590 alleles); SAS 0/22480 alleles.
BS1 strong Benign
Met: the same 2.78% non-cancer gnomAD popmax filtering AF far exceeds the APC VCEP BS1 threshold of 0.001%, though subsumed by BA1.
APC VCEP specification v2.1 (InSiGHT) BS1 (Benign Strong) rule: 'GnomAD Popmax Filtering Allele Frequency (AF) >= 0.001% (0.00001)', preferably based on the Popmax Filtering Allele Frequency of the non-cancer dataset from gnomAD (v2.1.1).Threshold derivation stated in the specification: prevalence 1:5,000, genetic heterogeneity 1, penetrance 0.8, allelic heterogeneity 0.06, calculated allele frequency >=0.0008%, rounded to >=0.001%; the calculated value equals the minor allele frequency of the most frequent pathogenic APC variant c.3927_3931delAAAGA p.(Glu1309Aspfs*4) on gnomAD v2.1.1.gnomAD v2.1 non-cancer exomes (VCEP-directed source, exome-only figures): popmax filtering AF 0.02778749 (2.78%), allele frequency 0.00186391 (249/133590 alleles) - roughly 2,780-fold and 186-fold above the 0.001% threshold respectively.
BS2 strong review Benign
Met: 18 homozygotes in gnomAD v3.1 non-cancer genomes and 33 in v4.1 exceed the APC VCEP >=2 homozygous-state requirement.
APC VCEP specification v2.1 (InSiGHT) BS2 (Benign Strong) rule: '>= 10 points for healthy individuals OR >= 2 times in homozygous state', with BS2_Supporting at '>= 3 points'; a healthy individual from control/non-cancer/normal/unaffected populations is worth 0.5 points.APC VCEP specification v2.1 (InSiGHT) BS2 instructions: the non-cancer dataset from gnomAD (v2.1.1) cannot be used for 'heterozygous healthy individuals' because of limited phenotype information and because it is usually already used for BA1/BS1, but it can be used to search for homozygous individuals.gnomAD v3.1 non-cancer genomes: 18 homozygotes (genome homozygote count 18; allele frequency 0.00856475, 1267/147932 alleles) - independently satisfies the >= 2 homozygous-state clause.
BP1 supporting Benign
Met at supporting: this missense lies at codon 26, outside the VCEP's excluded beta-catenin repeat (codons 1021-1035), in a gene where over 90% of pathogenic variants truncate.
APC VCEP CSPEC BP1 rule (only Supporting strength is applicable): 'BP1 is applicable to APC with the exception of missense variants located in the first 15-amino acid repeat of the beta-catenin binding domain (codon 1021-1035).'APC VCEP supplementary material (page 7, BP1 row): 'APC is a gene for which primarily truncating variants are known to cause disease. Based on our knowledge there are only two amino acid positions (1026 and 1028), where reported missense variants can be classified as Likely Pathogenic based on the APC-specific criteria ... Therefore, for all variants located in the first 15-amino acid repeat of the beta-catenin binding domain (codon 1021-1035) BP1 is not allowed to be used. A number of assumed "missense" variants are in fact splice variants. At least several splice prediction tools should be used.'The variant under assessment is p.(Ser26Arg) (NM_001127511.2:c.78C>A), a missense substitution at codon 26; codon 26 is far outside the BP1 exclusion window codons 1021-1035, so BP1 is allowed for this variant.
Assessed · not applied · 6 not met · 8 not assessed
Pathogenic
PVS1 Not met: c.78C>A is the missense change p.(Ser26Arg), not an APC null variant, so no PVS1 strength tier applies.
PS1 Not met: APC-specific PS1 applies only to the two Likely Pathogenic missense residues, 1026 and 1028; this variant changes Ser26, where no pathogenic missense is established.
PS2 Not assessed: no proband or parental testing exists in this case, so no APC VCEP de novo score (1-1.5 for PS2_Moderate) can be counted.
PS3 Not assessed: no functional assay exists for p.(Ser26Arg), and the VCEP protein-assay route covers only codons 959-2129, not codon 26.
PS4 Not assessed: no proband phenotype or family history was available to curate APC VCEP Table 1 phenotype points, and no case-control enrichment exists for this variant.
PM2 Not met: non-cancer gnomAD v2.1.1 allele frequency 0.186% is roughly 620-fold above the APC VCEP PM2 threshold of 0.0003%.
PM5 Not met: APC-specific PM5 is restricted to residues 1026 and 1028, and no same-residue missense comparator exists at Ser26.
PM6 Not assessed: no assumed-de-novo observation or parentage data are recorded, so no APC VCEP de novo score (0.5 for PM6_Supporting) can be counted.
PP1 Not assessed: zero genotyped relatives and zero meioses are documented, versus the APC VCEP Supporting threshold of 3-4 meioses in one family.
PP3 Not met: SpliceAI max delta 0.007 falls below the 0.2 supporting threshold, predicting no deleterious splice effect.
Benign
BS3 Not assessed: no RNA or beta-catenin-activity assay exists, and VCEP BS3 evidence is limited to synonymous/intronic RNA assays or codons 959-2129.
BS4 Not assessed: no affected relative lacking the variant is documented, so the APC VCEP BS4 phenotype-point thresholds (0.5-1 point) cannot be applied.
BP2 Not met: no report of c.78C>A in trans with a pathogenic APC variant and zero unknown-phase co-occurrences versus the required three.
BP5 Not assessed: no colorectal polyposis phenotype record and no other-gene testing results were available to establish or exclude an alternate genetic basis.
N/A · 10 PM1 · PM3 · PM4 · PP2 · PP4 · PP5 · BP3 · BP4 · BP6 · BP7
Research & evidence
Population frequency · supports benign
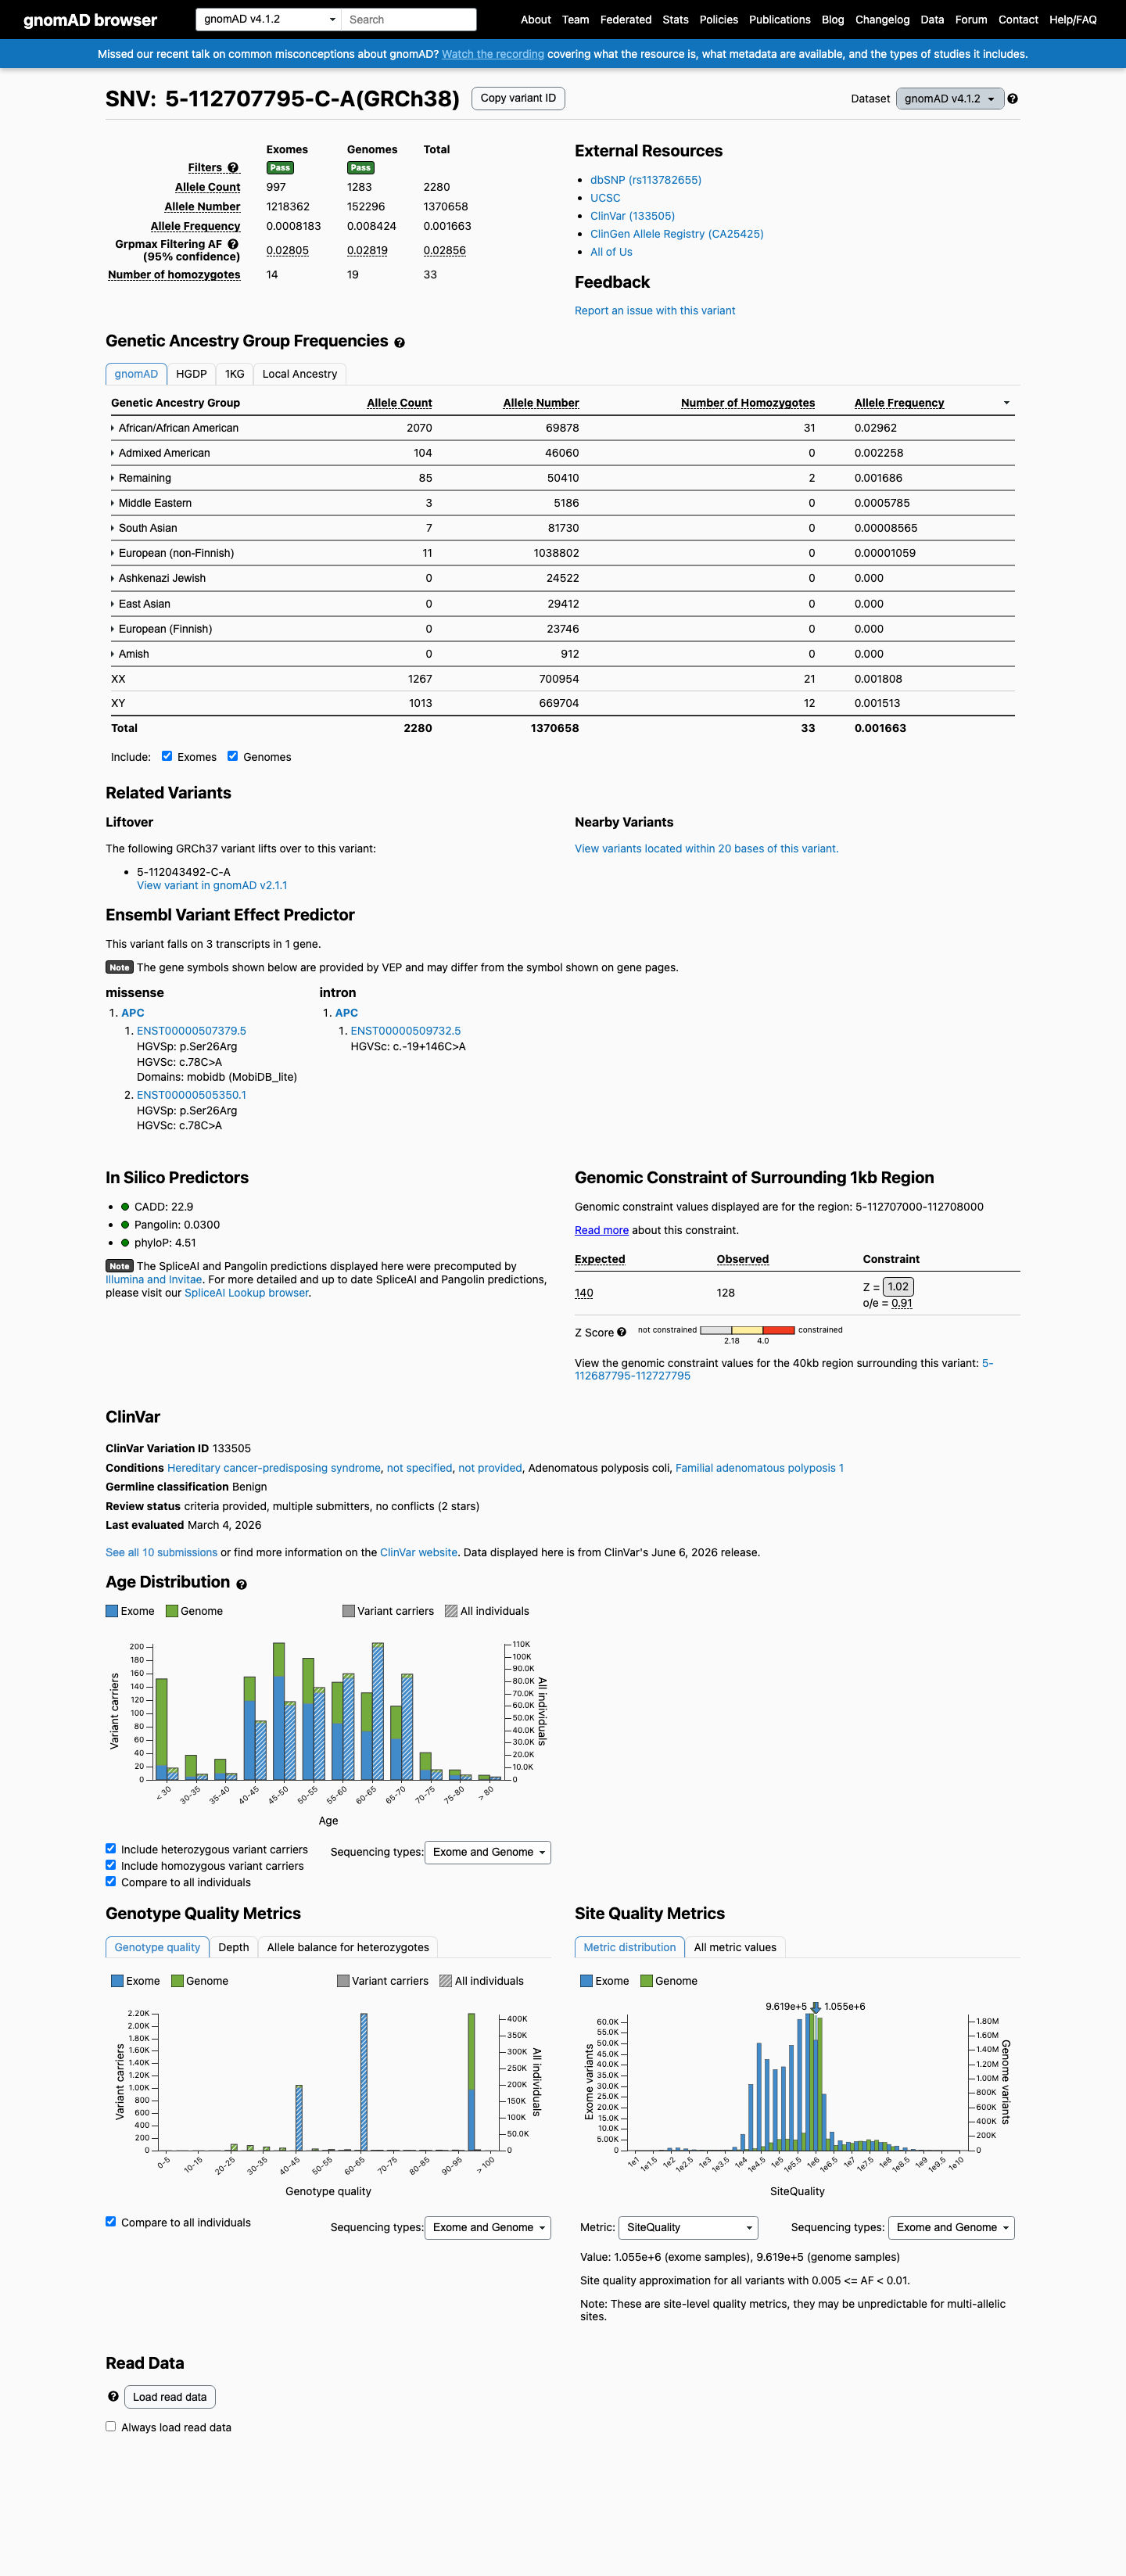
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
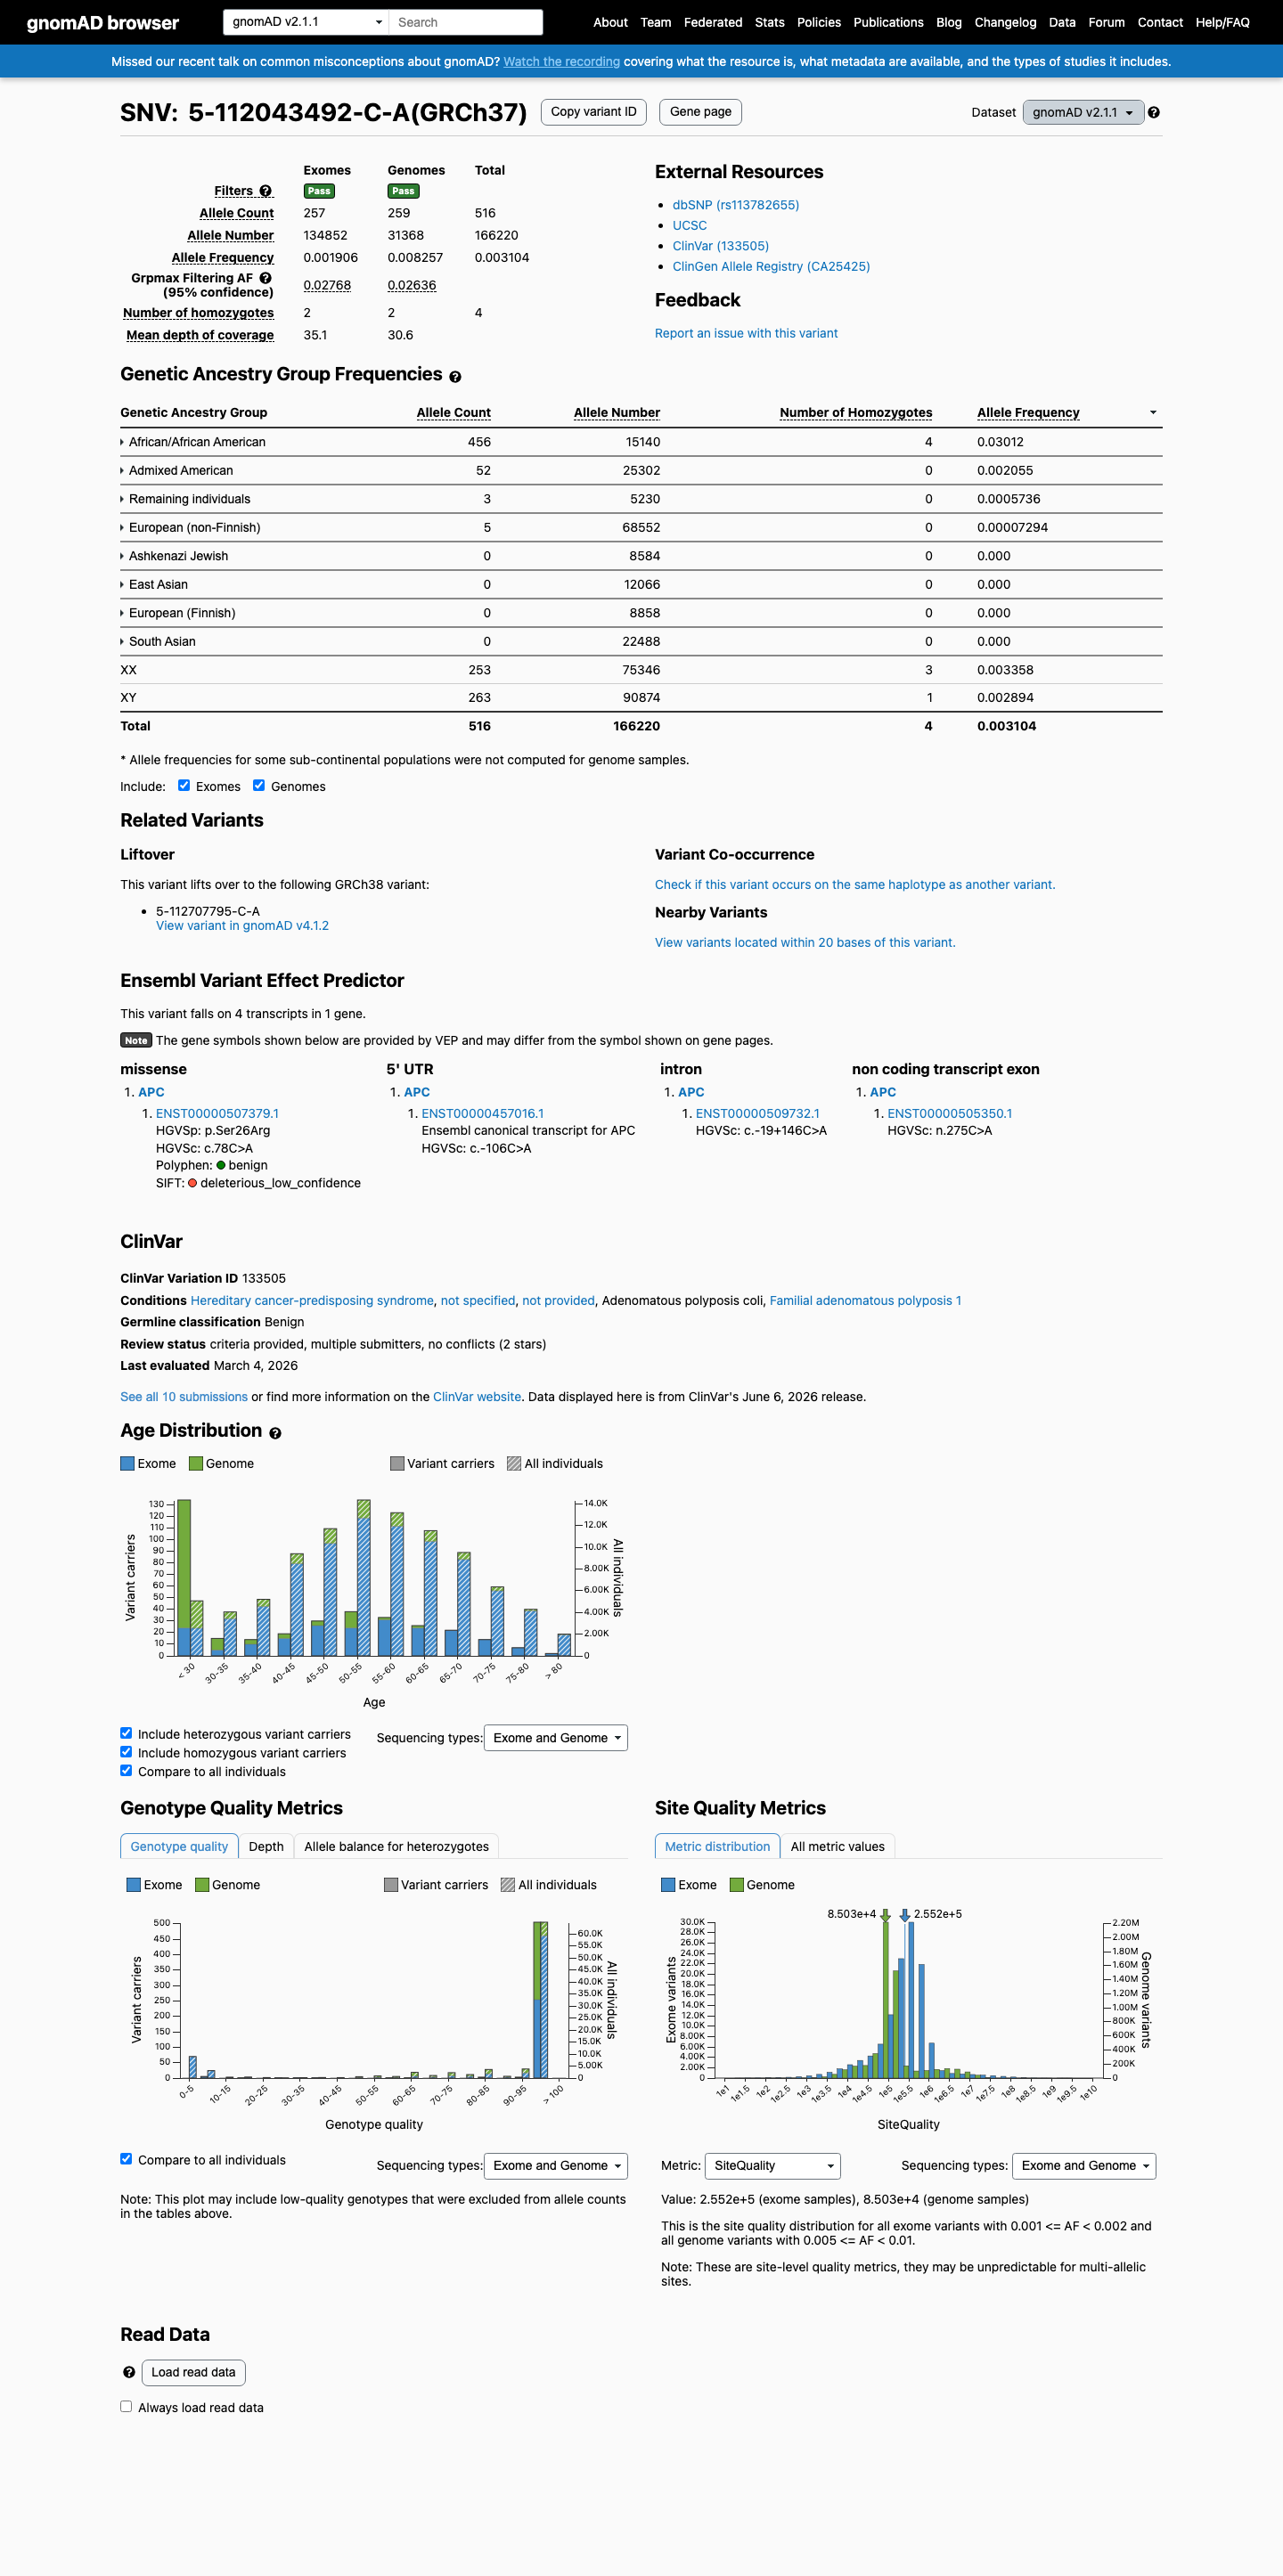
This variant is present in gnomAD v4.1 (AF= 0.00166343; MAF= 0.16634%, 2280/1370658 alleles, homozygotes = 33) and has highest observed frequency in the African/African American population (AF= 0.0296231; MAF= 2.96231%, 2070/69878 alleles, homozygotes = 31); grpmax FAF= 0.0285595.
v2.1
This variant is present in gnomAD v2.1 (AF= 0.00310432; MAF= 0.31043%, 516/166220 alleles, homozygotes = 4) and has highest observed frequency in the African/African American population (AF= 0.0301189; MAF= 3.01189%, 456/15140 alleles, homozygotes = 4); grpmax FAF= 0.02768.
🇨🇦 CA
This variant is present in gnomAD-Canada v1.0 (AF= 0.001956521739130435, 36/18400 alleles, homozygotes = 2).
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.17% · 2280 / 1,370,658
33 hom · FAF 2.9%
African/African American
2070 / 69,878
3%
31 hom
Admixed American
104 / 46,060
0.23%
Remaining individuals
85 / 50,410
0.17%
2 hom
Middle Eastern
3 / 5,186
0.058%
South Asian
7 / 81,730
0.0086%
European (non-Finnish)
11 / 1,038,802
0.0011%
+ 4 not observed (European (Finnish), Amish, East Asian, Ashkenazi Jewish)
gnomAD v2.1
0.31% · 516 / 166,220
4 hom · FAF 2.8%
African/African American
456 / 15,140
3%
4 hom
Admixed American
52 / 25,302
0.21%
Remaining individuals
3 / 5,230
0.057%
European (non-Finnish)
5 / 68,552
0.0073%
+ 4 not observed (Ashkenazi Jewish, East Asian, European (Finnish), South Asian)
gnomAD Canada 🇨🇦
0.2% · 36 / 18,400
2 hom · FAF 2.5%
African/African American
35 / 1,018
3.4%
2 hom
Latino/Admixed American
1 / 836
0.12%
+ 7 not observed (Ashkenazi Jewish, East Asian, European (Finnish), Middle Eastern, European (non-Finnish), Remaining individuals, South Asian)
ClinVar screenshot
ClinVar
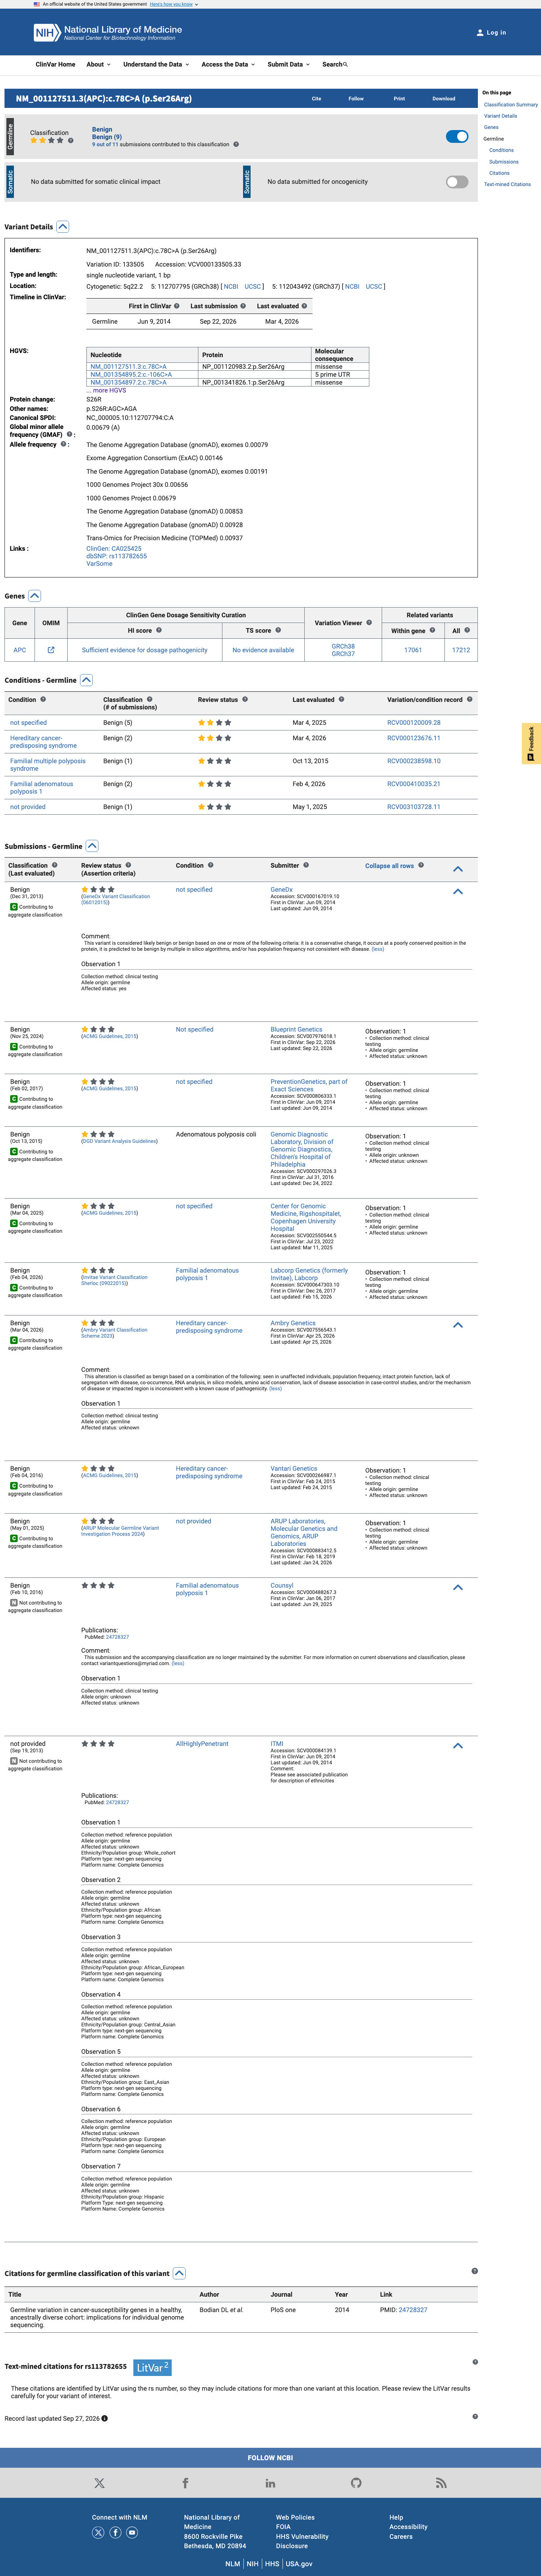
This variant has been reported in ClinVar as Benign (10 clinical laboratories). (ClinVarID = 133505)
SpliceAI screenshot
In silico
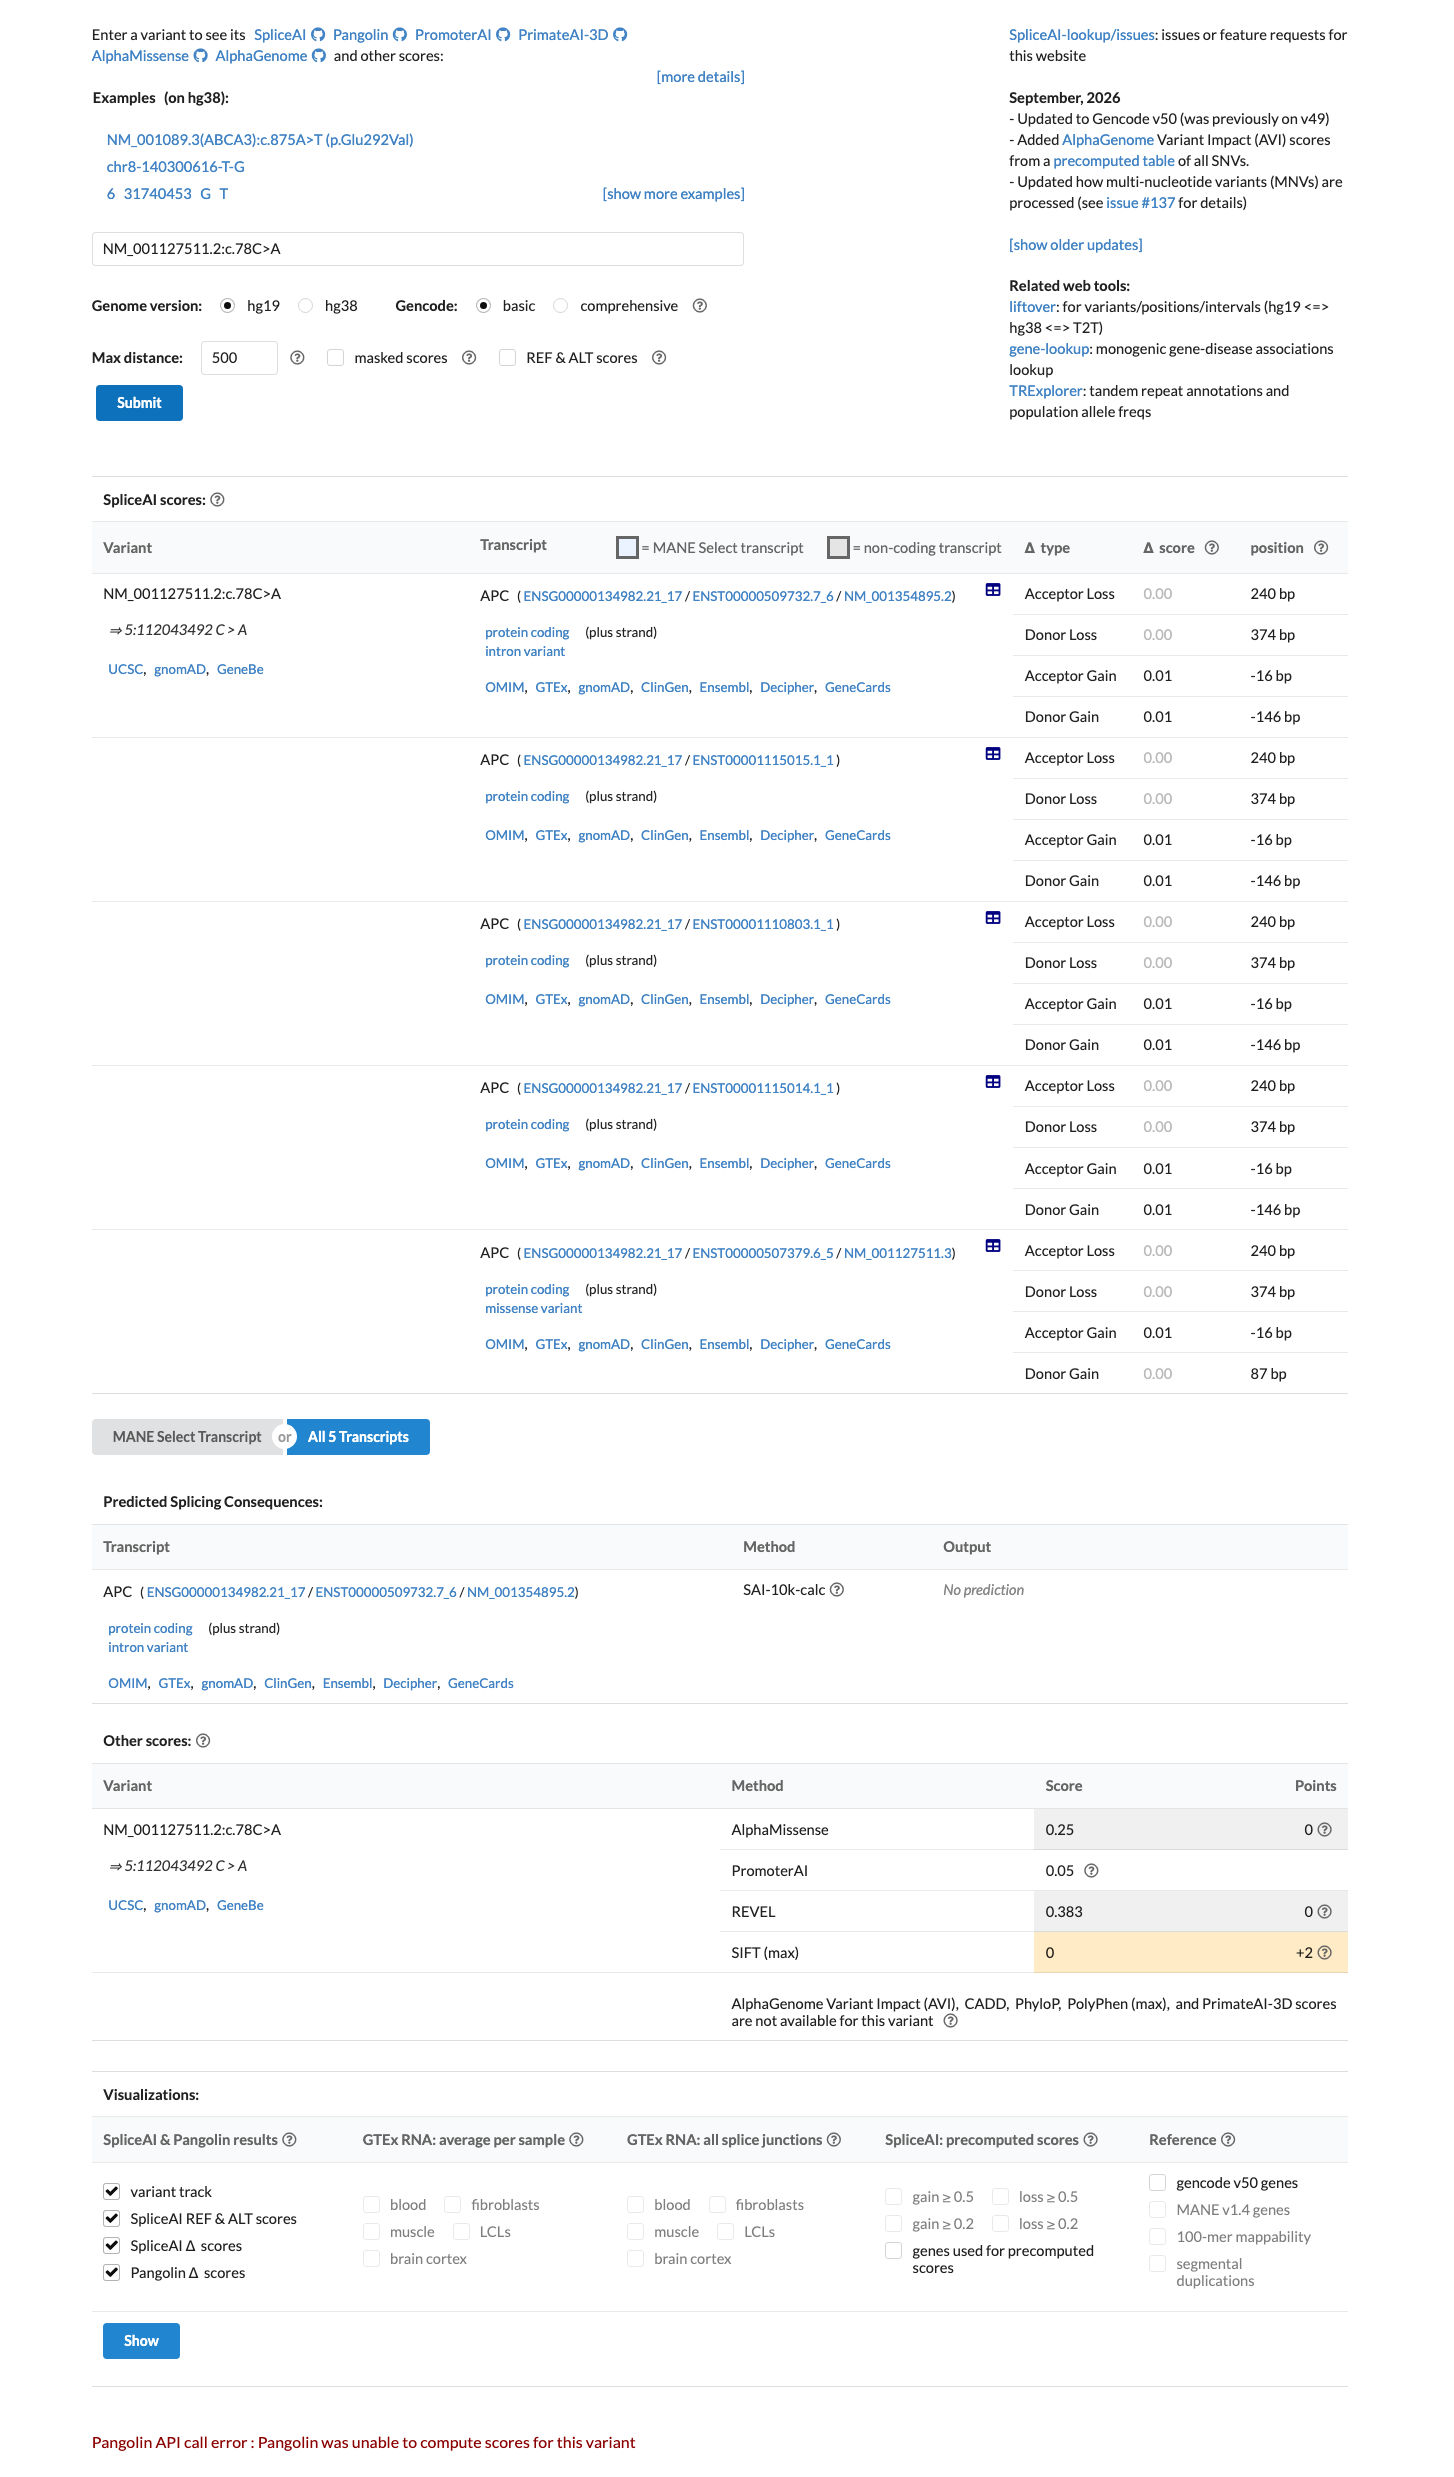
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.01). REVEL score = 0.383. BayesDel score = -0.0215517.
Functional / OncoKB screenshot
Functional
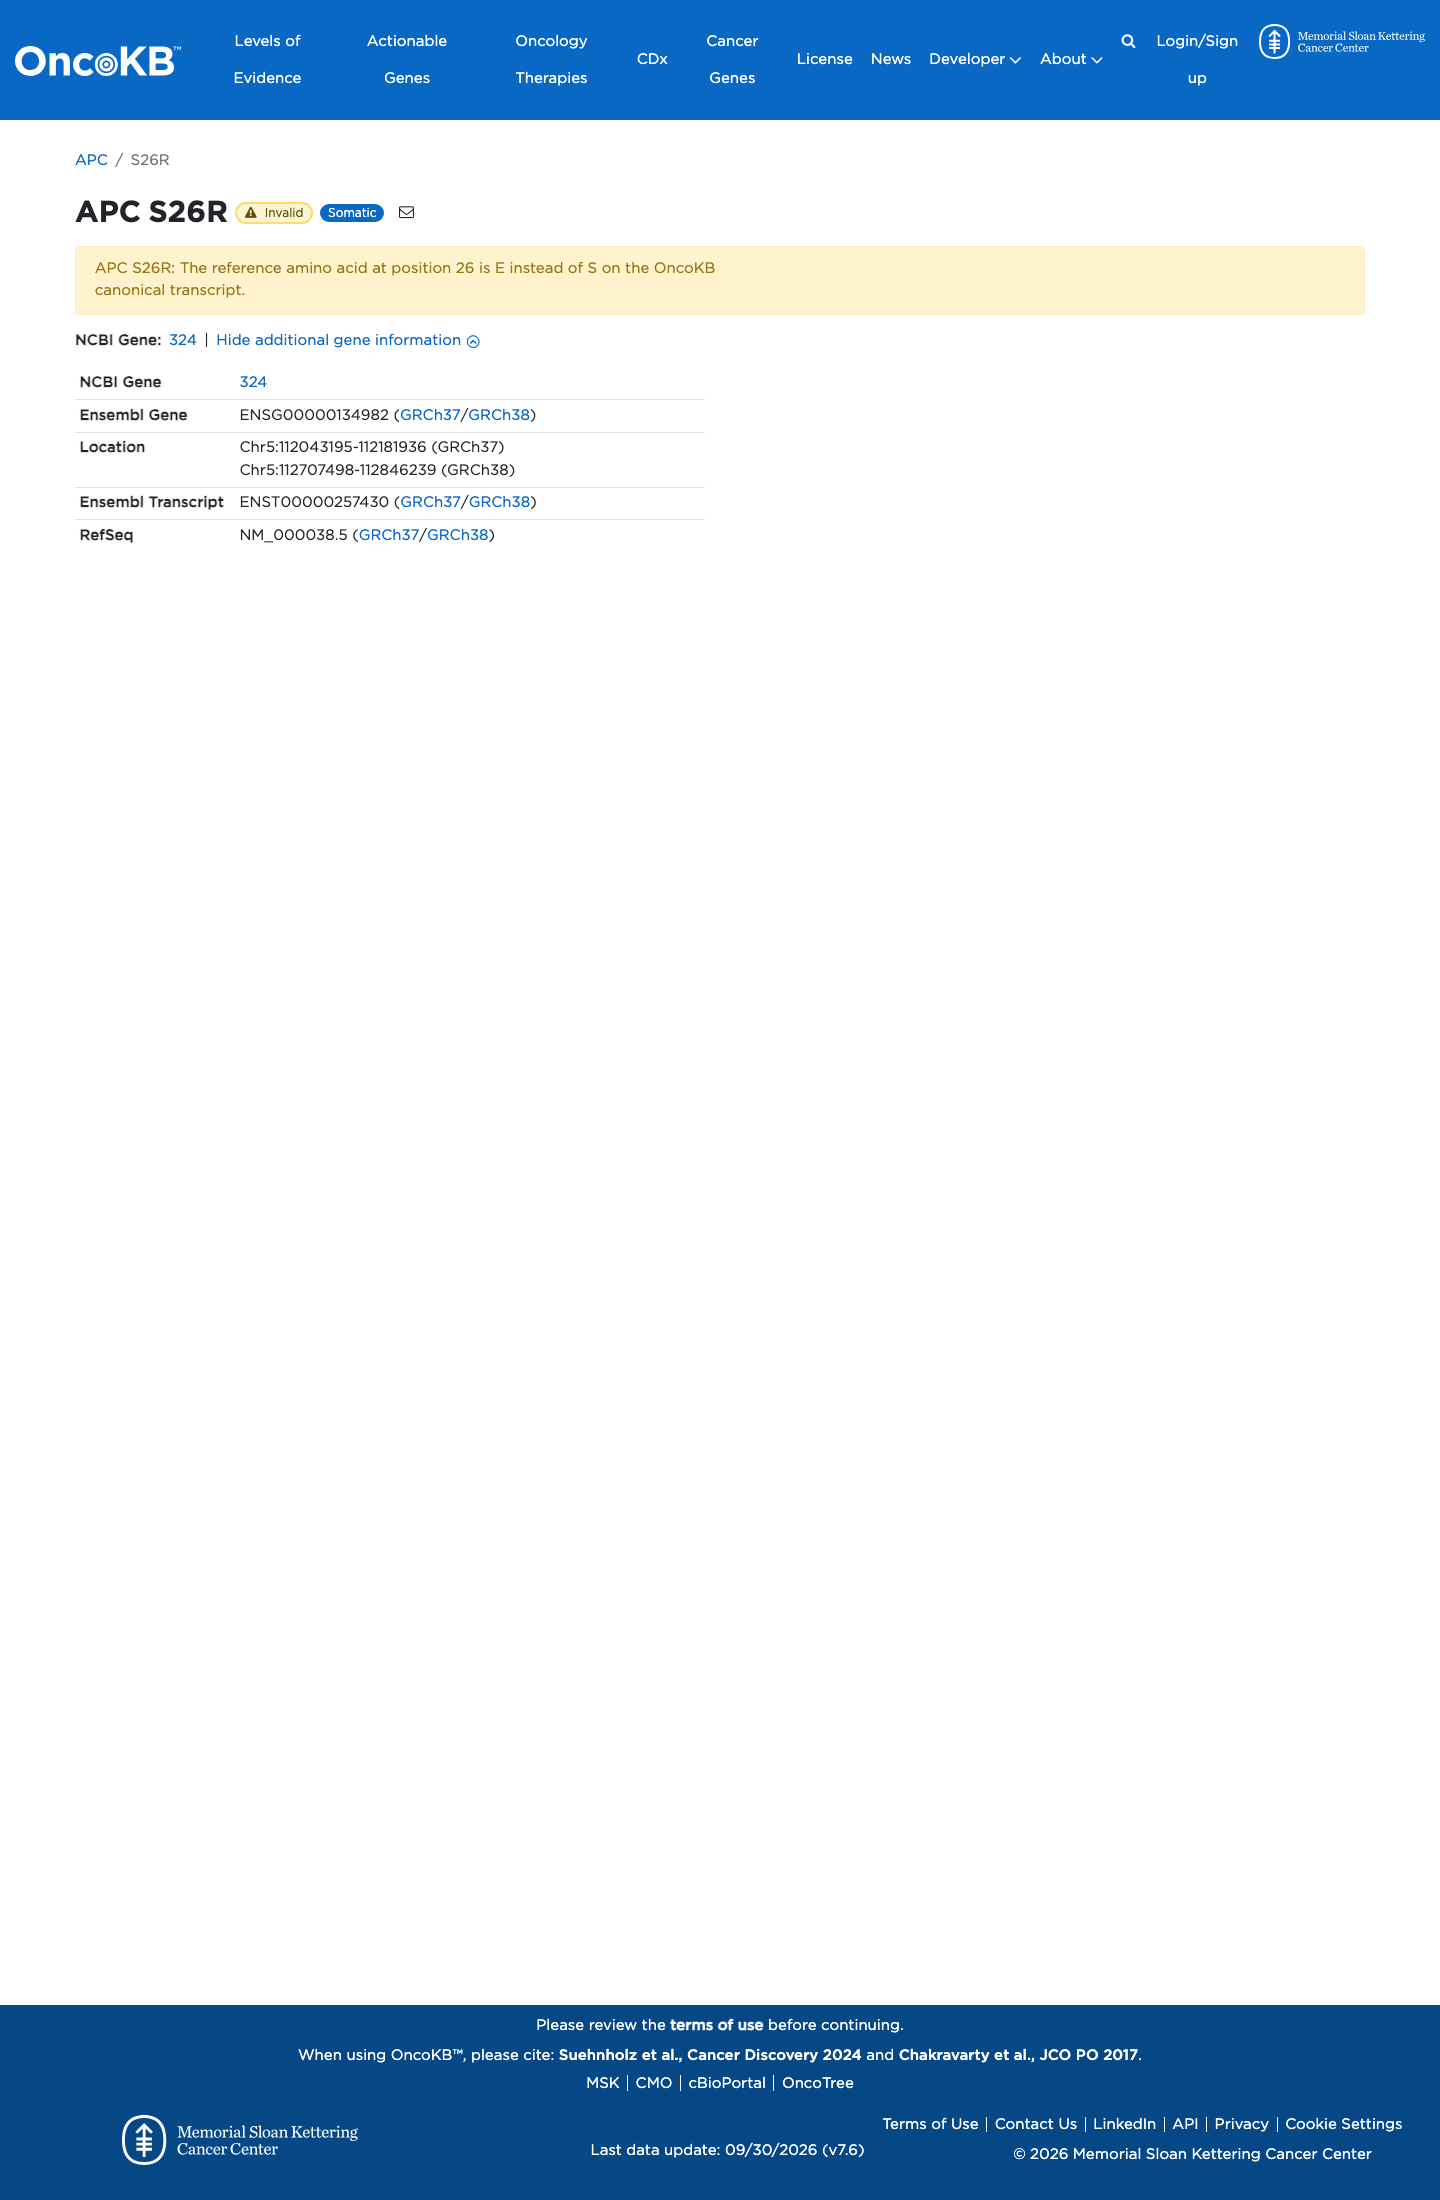
OncoKB identified curated literature and non-variant-specific oncogenicity context for review; listed oncogenicity label: not classified.
OncoKB ↗
COSMIC screenshot
COSMIC
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
4papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 4 further PMIDs triaged but not cited — see Sources & references.
Rule & framework references · cited for criterion definitions, not variant evidence
21368914 ↗ Clinical utility gene card for: familial adenomatous polyposis (FAP) and attenuated FAP (AFAP).
24728327 ↗ Germline variation in cancer-susceptibility genes in a healthy, ancestrally diverse cohort: implications for individual genome sequencing.
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint co
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification crite
Sources & reference links
9Sources
CSpec VCEP
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 4 PMIDs not cited in assessment
15604628 ↗ Genetic cancer risk assessment and counseling: recommendations of the national society of genetic counselors. CLINVAR
20301519 ↗ APC-Associated Polyposis Conditions. CLINVAR
25645574 ↗ ACG clinical guideline: Genetic testing and management of hereditary gastrointes CLINVAR
26389505 ↗ Genetics of Colorectal Cancer (PDQ®): Health Professional Version. CLINVAR