Back
NM_001128425.2:c.505G>A
p.Glu169Lys · MUTYH
0%
complete
Final classification
VUS
PM2PP3
MUTYH
c.505G>A
p.Glu169Lys
missense · exon 7

MUTYH encodes a DNA repair enzyme (a glycosylase) that fixes oxidative DNA damage by removing adenine bases that have been mistakenly paired with guanine or with oxidized guanine lesions. Inherited mutations in both copies of the gene cause MUTYH-associated polyposis (MAP), a recessive condition that strongly predisposes people to multiple colorectal polyps and colorectal cancer. The gene acts as a tumor suppressor whose loss of function allows DNA damage to accumulate, and somatic changes in it have also been reported in colon cancer, though whether they drive cancer on their own is not fully established.

This variant

MUTYH encodes a DNA-repair glycosylase, and pathogenic variants in both gene copies cause recessive MUTYH-associated polyposis with increased colorectal polyp and cancer risk.

Transcript
NM_001128425.2
HGVS · transcript:coding
NM_001128425.2:c.505G>A
GRCh38
chr1:45332834 C>T
GRCh37
chr1:45798506 C>T
VUS: PM2 (supporting) plus PP3 (moderate) do not meet generic ACMG/AMP thresholds for pathogenic, likely pathogenic, likely benign, or benign classification.
Classification rationale
PM2PP3 VUS
MUTYH c.505G>A missense · exon 7

PM2 supporting: gnomAD v4.1 total allele frequency is 6.19e-7, below 0.0001. PP3 moderate: REVEL 0.846 exceeds the calibrated moderate threshold of 0.773.

PM2 + PP3 → VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_001128425.2 · variants mapped to exon structure
MUTYH NM_001128425.2
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 22 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
PM2 supporting Pathogenic
Met at supporting: gnomAD v4.1 total AF is 6.19e-7, below the 0.0001 PM2 threshold, with zero homozygotes.
The InSiGHT MUTYH specification is available, but its supplied ruleset has no criterion-specific population threshold; the supplied generic PM2 default was used.gnomAD v4.1 reports total AF 6.194911747287248e-07 from 1 alternate allele among 1,614,228 alleles, zero homozygotes, and highest subpopulation AF 1.5997440409534473e-05.The variant is absent from gnomAD v2.1 and from the reported non-cancer gnomAD v2.1 and v3.1 subsets.
PP3 moderate Pathogenic
Met (moderate): REVEL 0.846 meets the published PP3 moderate threshold of >=0.773.
Variant normalization identifies NM_001128425.2:c.505G>A as the missense substitution NP_001121897.1:p.(Glu169Lys).The local REVEL lookup reports a score of 0.846 for this variant.The published ClinGen SVI REVEL calibration assigns moderate PP3 at REVEL >=0.773 (Pejaver et al. 2022, PMID:36413997).
Assessed · not applied · 6 not met · 16 not assessed
Pathogenic
PS1 Not assessed: no established pathogenic p.Glu169Lys comparator was identified, and ClinVar classifies the target variant as uncertain significance.
PS2 Not assessed: no documented parental genotypes or confirmed de novo observation is available for MUTYH c.505G>A.
PS3 Not assessed: no validated variant-specific functional assay or damaging biological readout was identified for p.Glu169Lys.
PS4 Not assessed: no exact-variant case-control counts or enrichment metric such as an odds ratio, relative risk, or p-value is available.
PM1 Not assessed: no approved MUTYH domain entry or statistically significant hotspot places residue 169 in a critical region.
PM3 Not assessed: no affected-proband observation or phase-resolved second MUTYH variant is documented, so PM3 cannot be assigned.
PM5 Not assessed: no validated pathogenic or likely pathogenic alternate missense variant at MUTYH residue 169 was identified.
PM6 Not assessed: no publication or case record reports c.505G>A as presumed de novo without parental testing.
PP1 Not assessed: no affected relatives, parental phase, biallelic genotypes, or informative meioses are reported for segregation analysis.
PP2 Not assessed: no validated MUTYH missense-enrichment estimate or applicable PP2 rule is available.
PP4 Not assessed: no patient phenotype, polyp burden, cancer history, or family history is documented to compare with MUTYH-associated polyposis.
PP5 Not met: exact-variant ClinVar classification is Uncertain significance with 0 expert-panel submissions, not Pathogenic or Likely pathogenic.
Benign
BA1 Not met: gnomAD v4.1 total AF is 6.19e-7, far below the 0.05 BA1 stand-alone threshold.
BS1 Not met: the highest gnomAD v4.1 subpopulation AF is 1.60e-5, below the generic 0.01 BS1 threshold.
BS2 Not met: gnomAD v4.1 reports zero homozygotes, so no unaffected biallelic individual supports recessive BS2.
BS3 Not assessed: no validated variant-specific functional assay or normal biological readout was identified for p.Glu169Lys.
BS4 Not assessed: no tested relatives or phenotype-genotype comparisons are available to evaluate non-segregation.
BP1 Not assessed: no validated evidence shows that MUTYH disease is predominantly caused by truncating rather than missense variants.
BP2 Not assessed: no validated cis/trans relationship between c.505G>A and another pathogenic variant is documented for a benign-evidence determination.
BP4 Not met: REVEL 0.846 exceeds the highest applicable BP4 threshold of <=0.29.
BP5 Not assessed: no patient phenotype or well-established alternate molecular diagnosis is available to support BP5.
BP6 Not met: exact-variant ClinVar classification is Uncertain significance with 0 expert-panel submissions, not Benign or Likely benign.
N/A · 4 PVS1 · PM4 · BP3 · BP7
Research & evidence
Population frequency
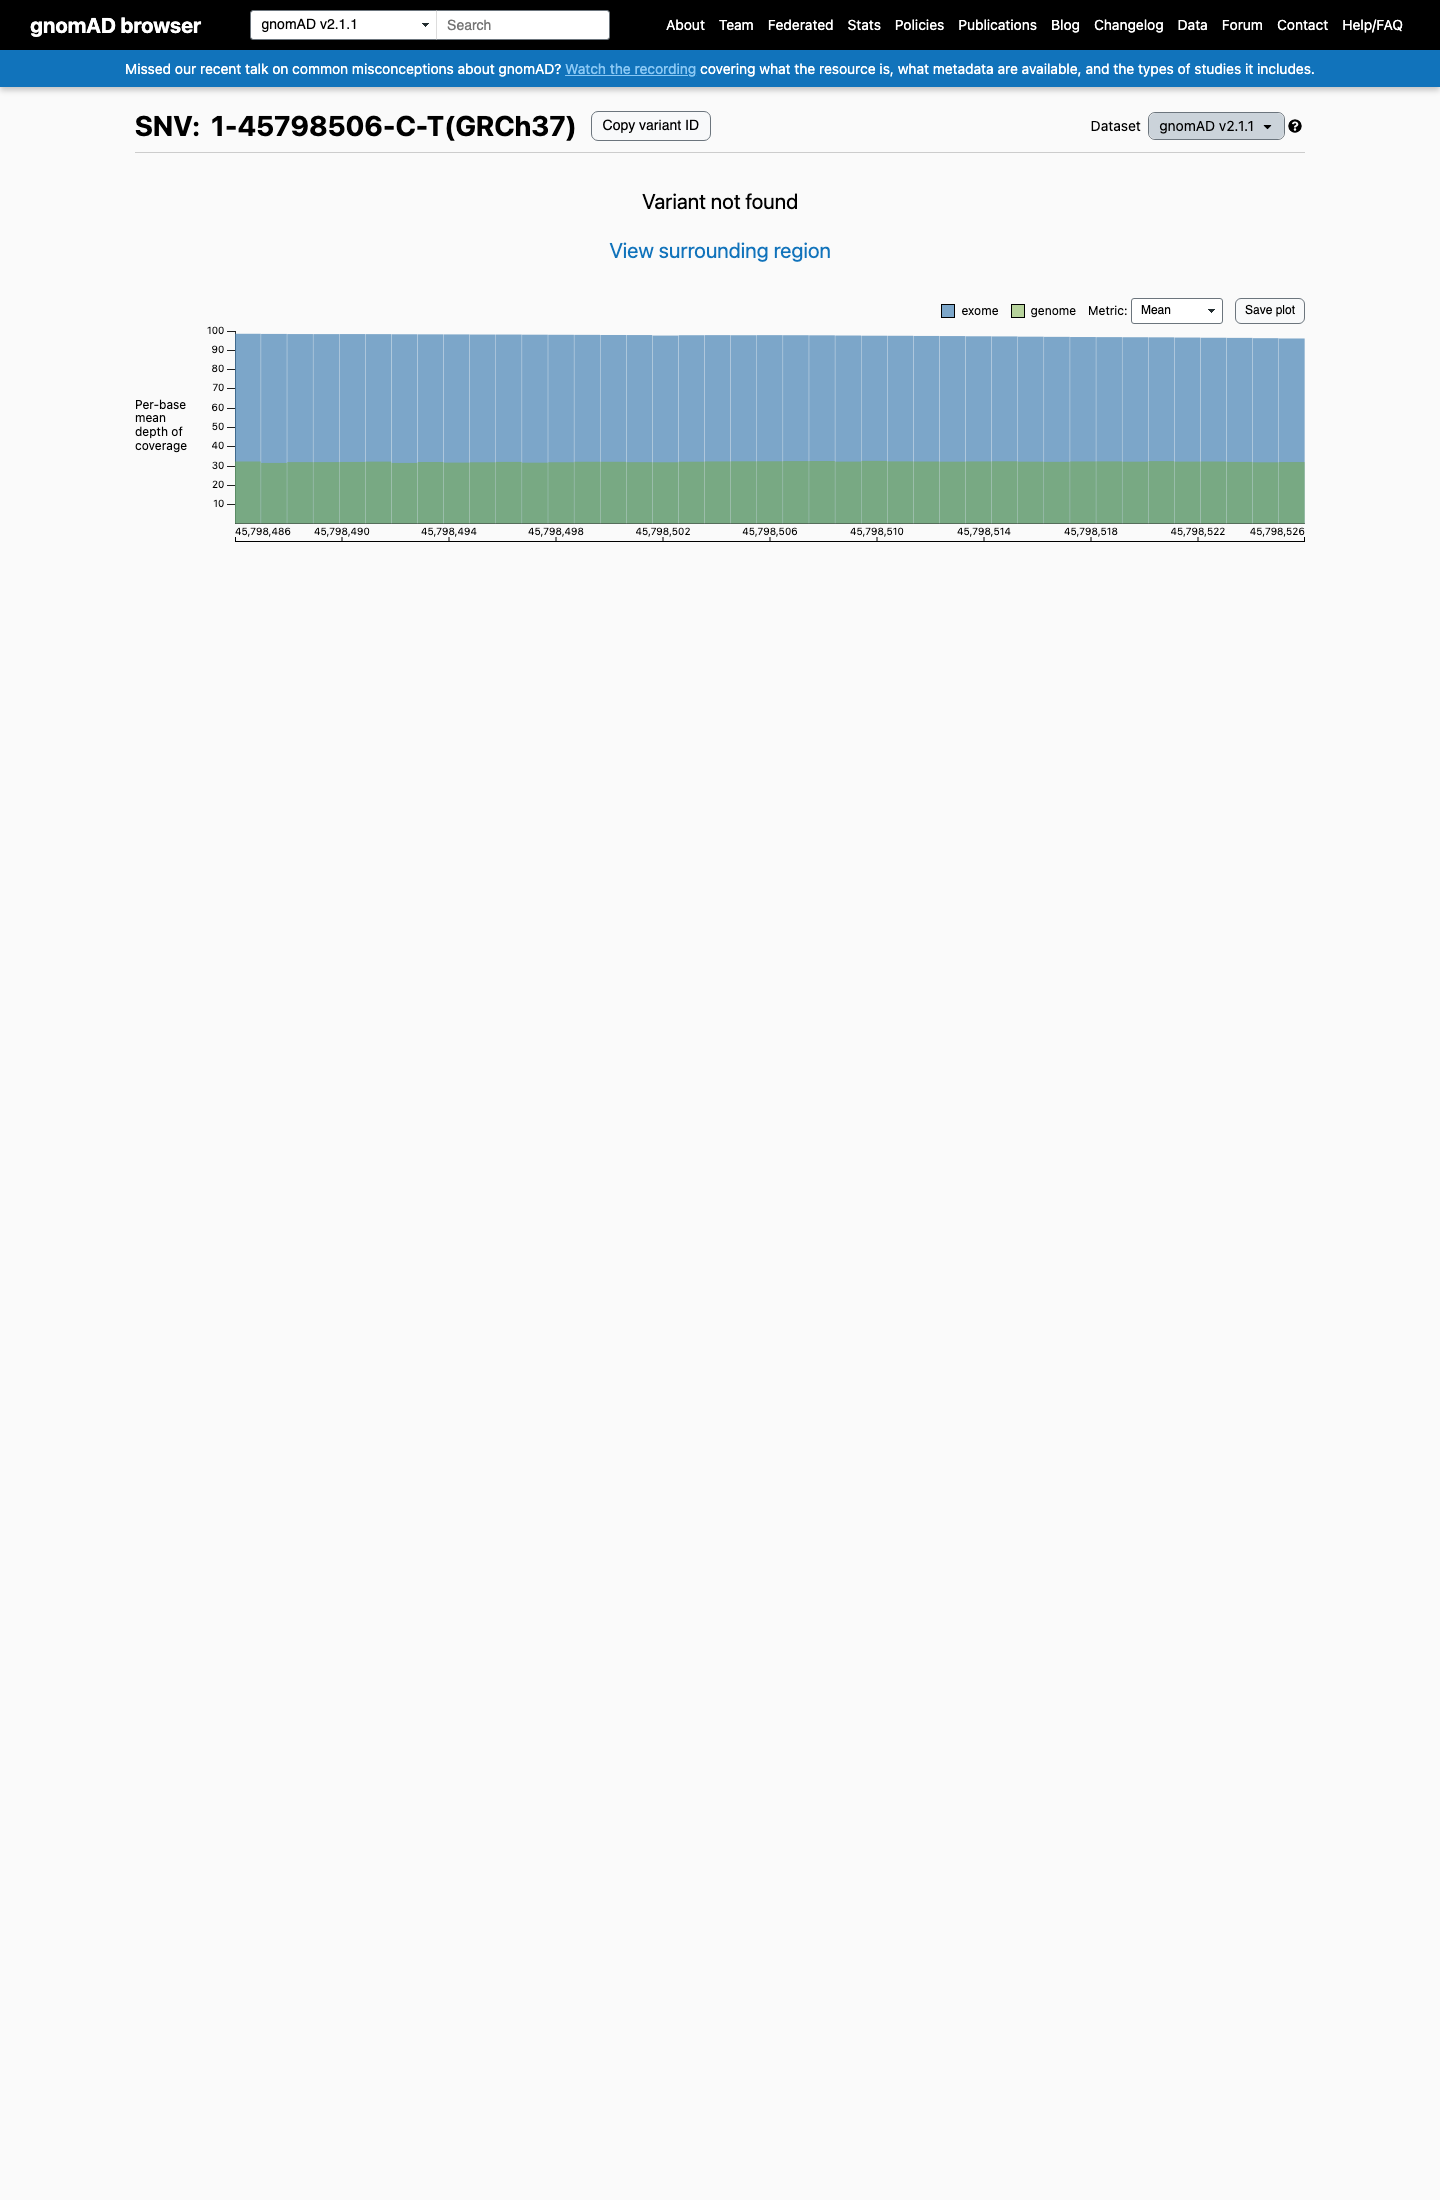
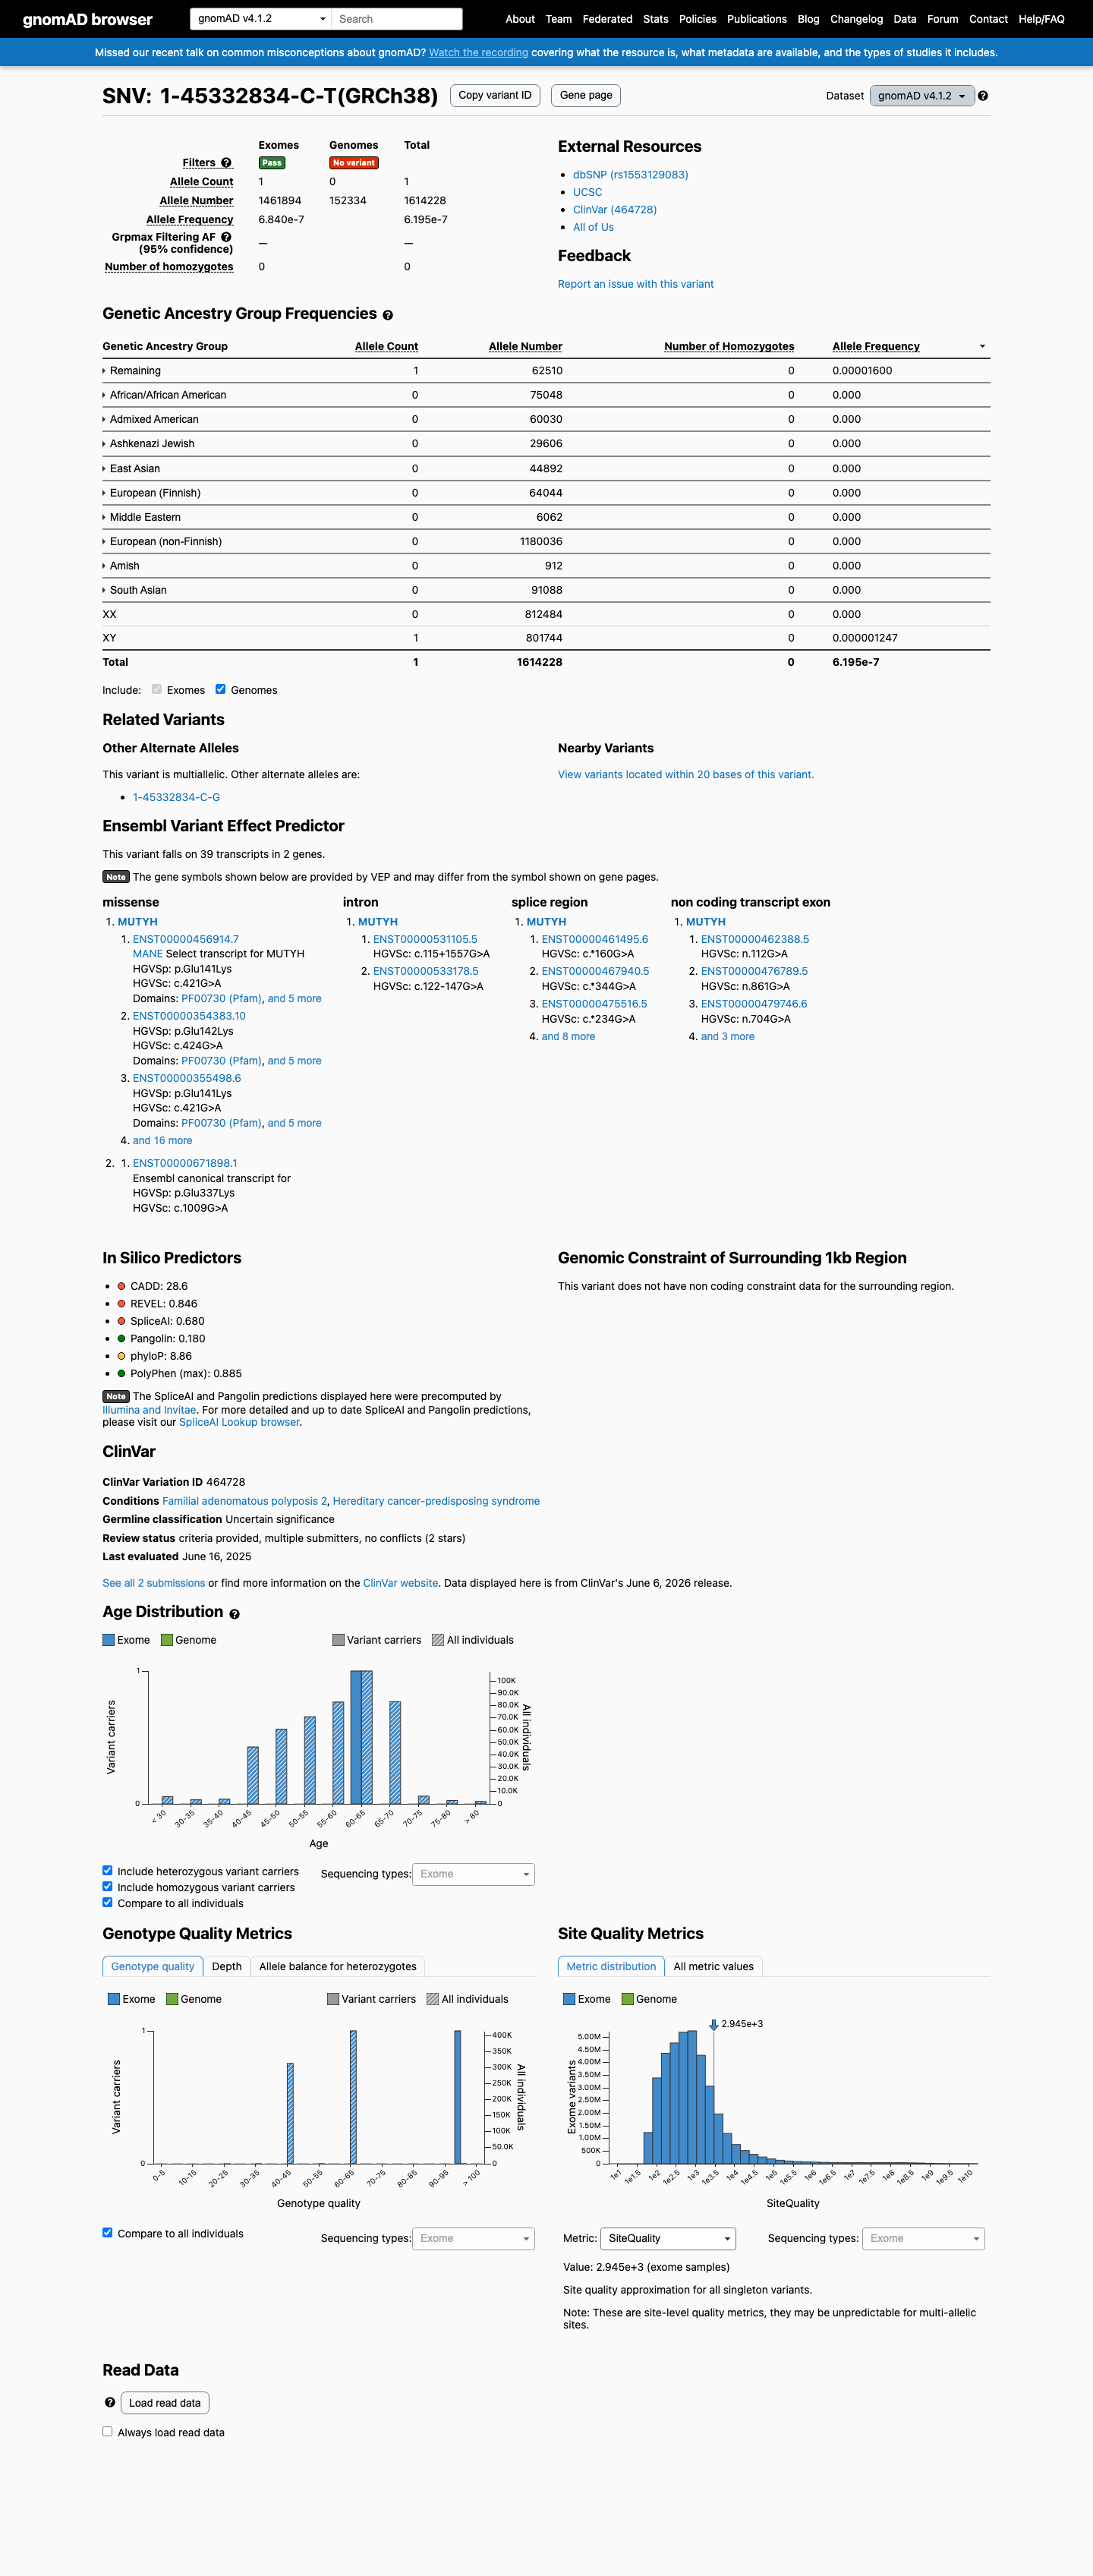
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 6.19491e-07; MAF= 0.00006%, 1/1614228 alleles, homozygotes = 0) and has highest observed frequency in the Remaining individuals population (AF= 1.59974e-05; MAF= 0.00160%, 1/62510 alleles, homozygotes = 0).
v2.1
Absent from gnomAD v2.1.
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
6.2e-05% · 1 / 1,614,228
0 hom
Remaining individuals
1 / 62,510
0.0016%
+ 9 not observed (Admixed American, European (Finnish), Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American, European (non-Finnish))
gnomAD v2.1
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
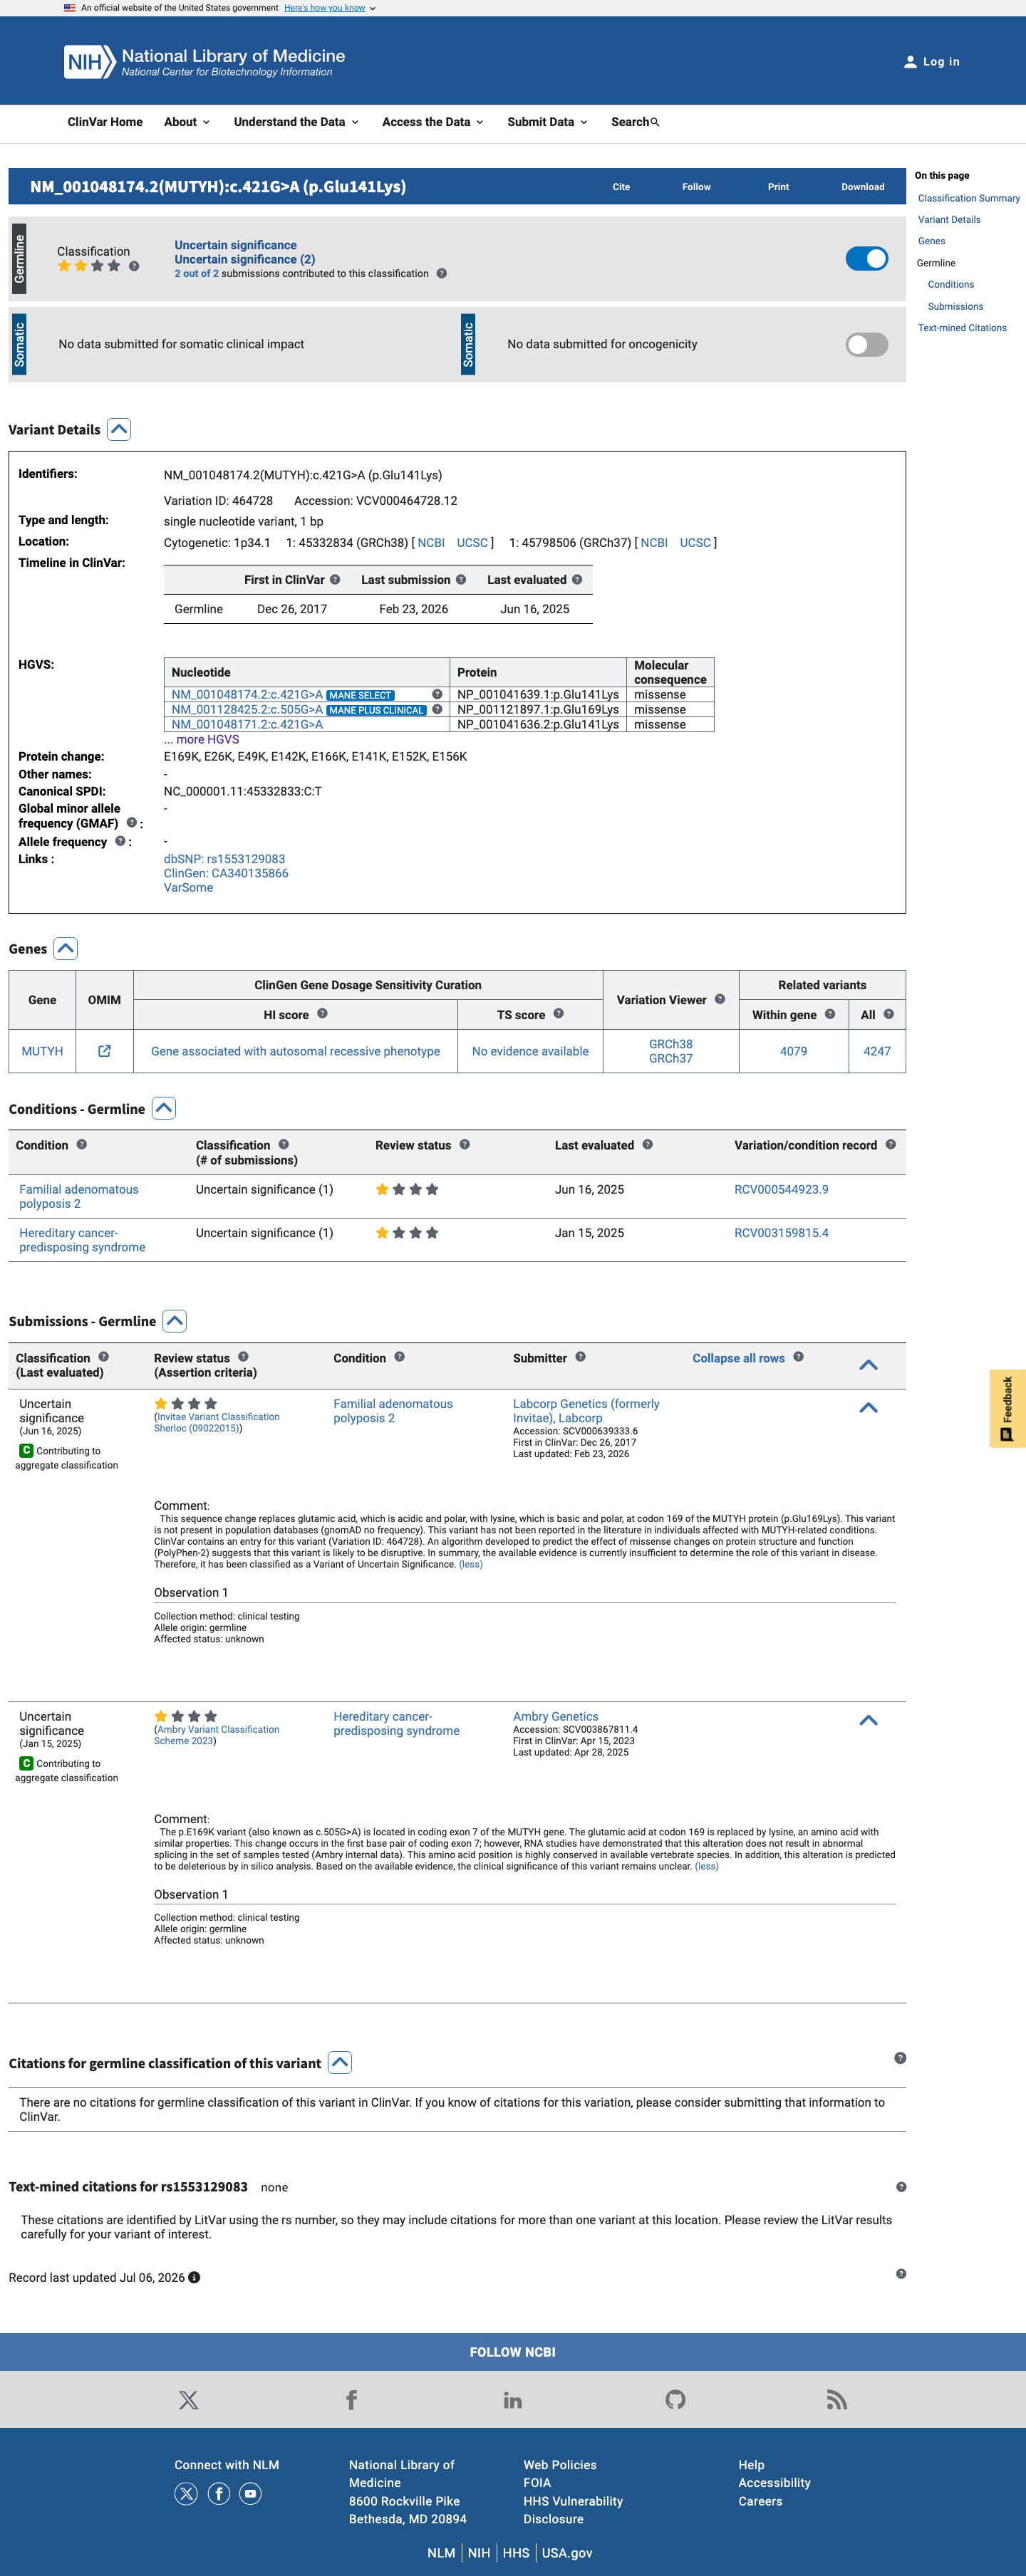
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Uncertain significance (2 clinical laboratories). (ClinVarID = 464728)
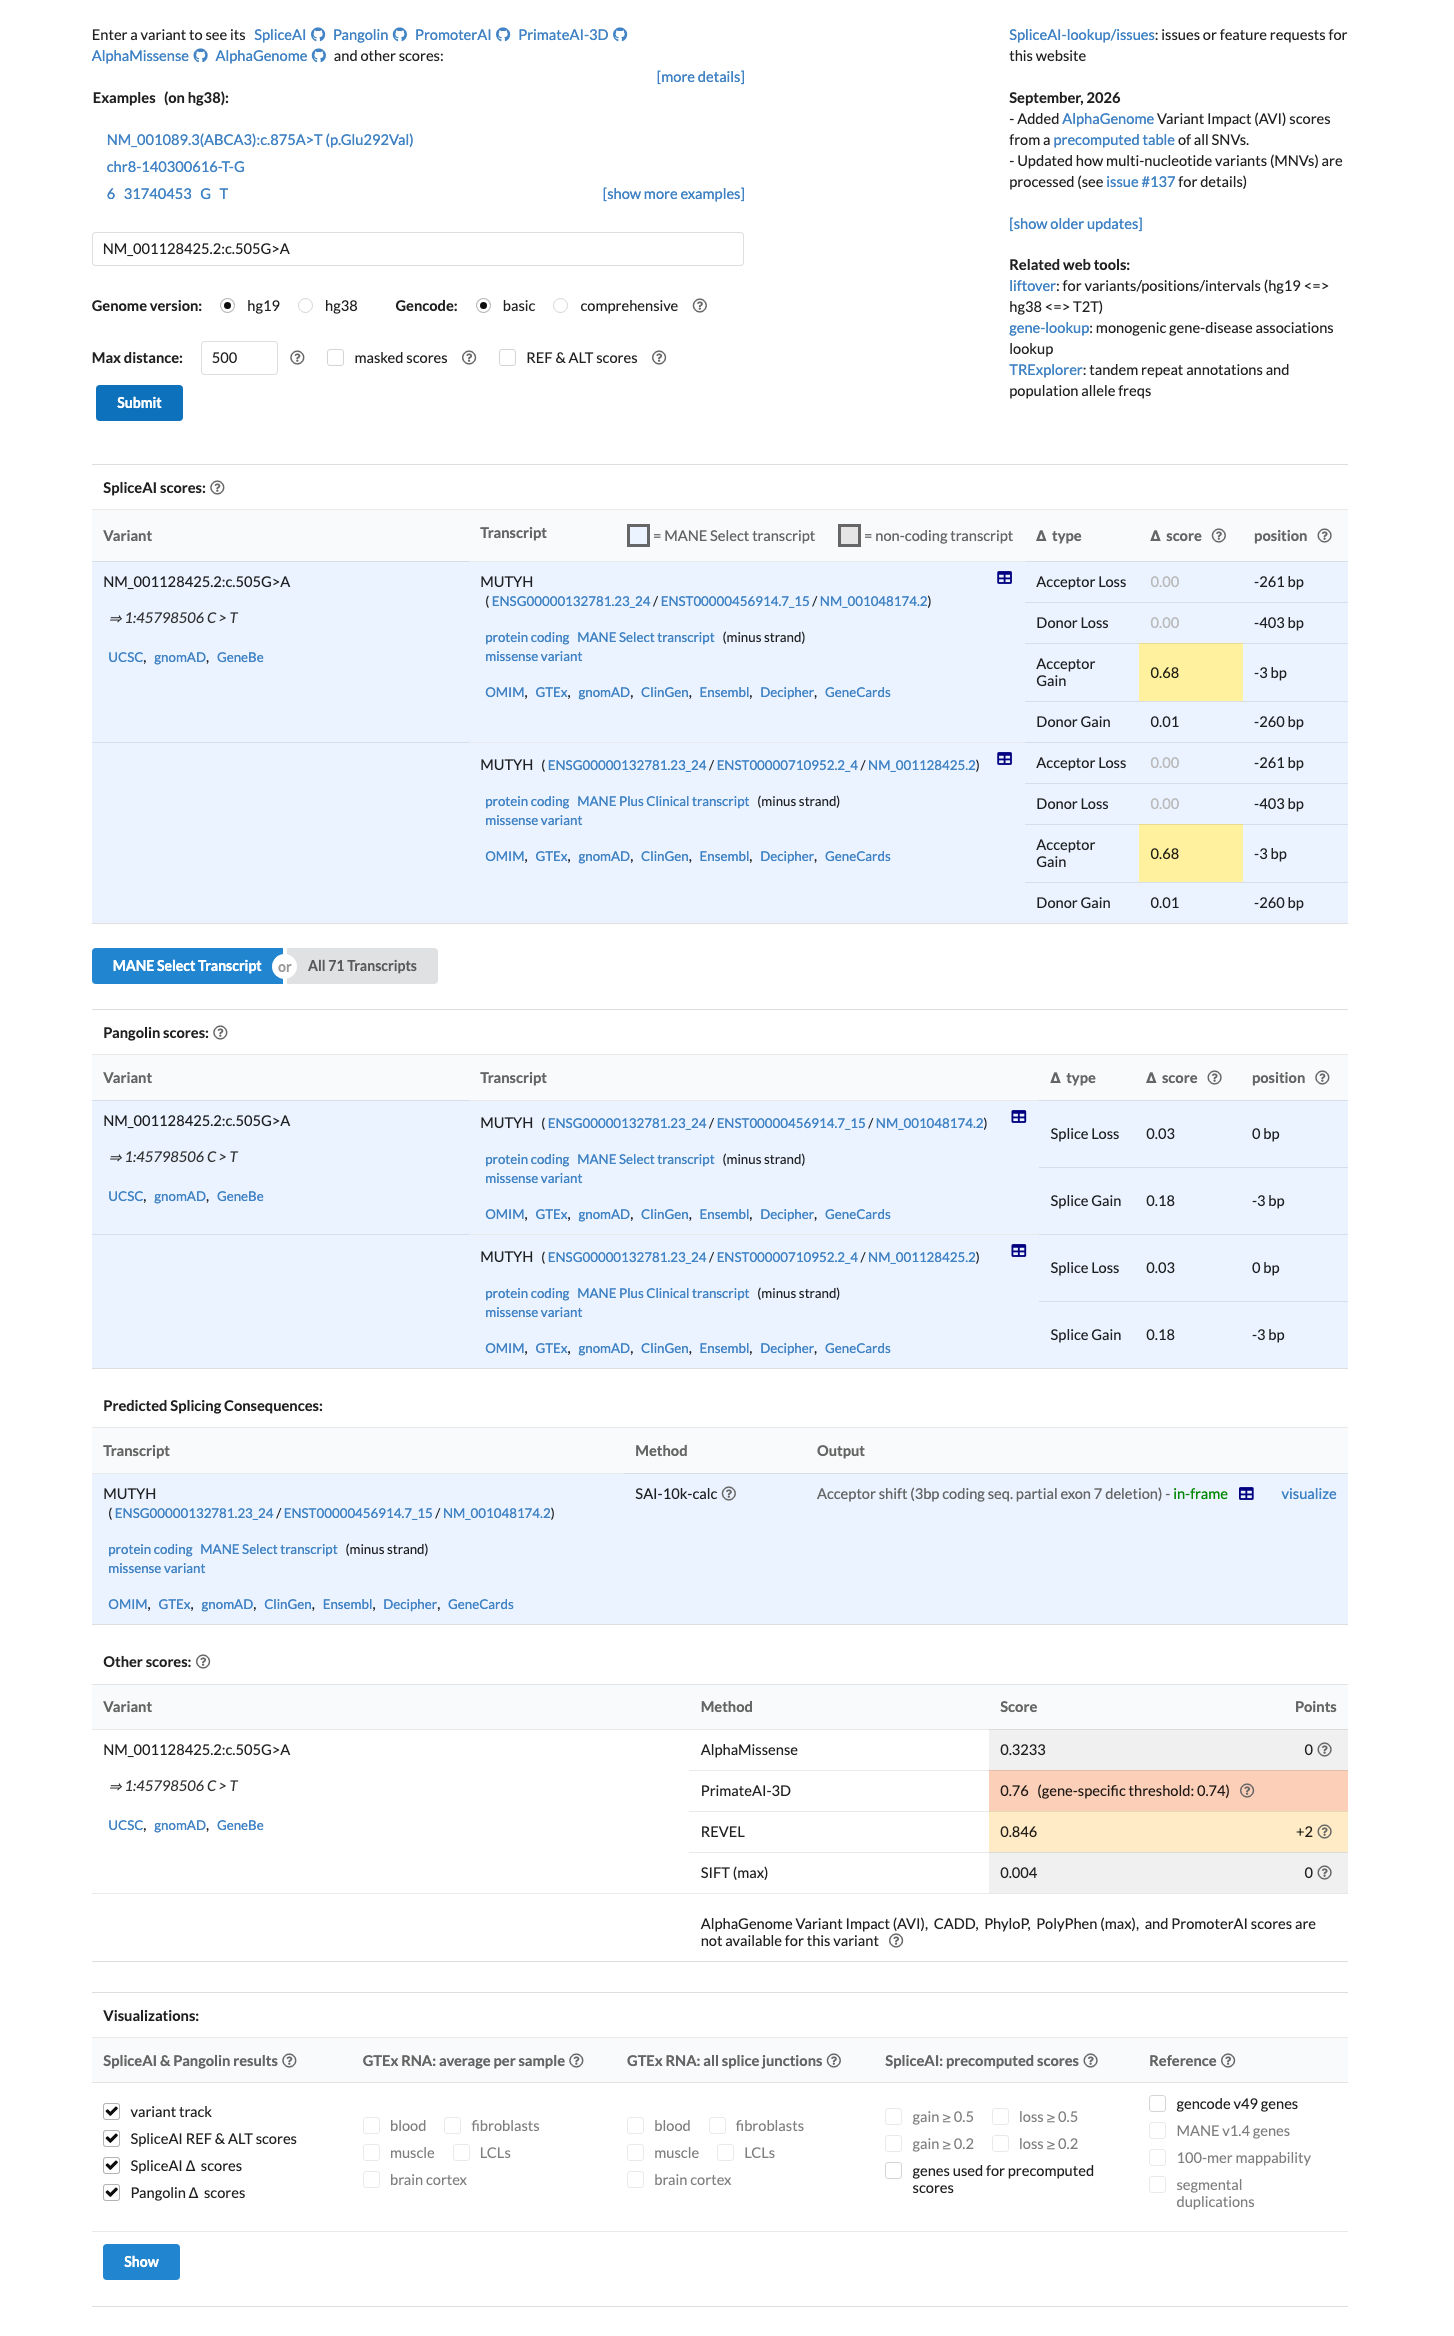
SpliceAI screenshot
In silico
SpliceAI returned NO scores for this variant, so no SpliceAI-based splice prediction is available. This is missing data, NOT evidence of absent splice impact: it must not be used to support BP4 or to argue against PP3/PVS1. Pangolin scores may be present but are not calibrated for PP3/BP4 here. REVEL score = 0.846. BayesDel score = 0.336123.
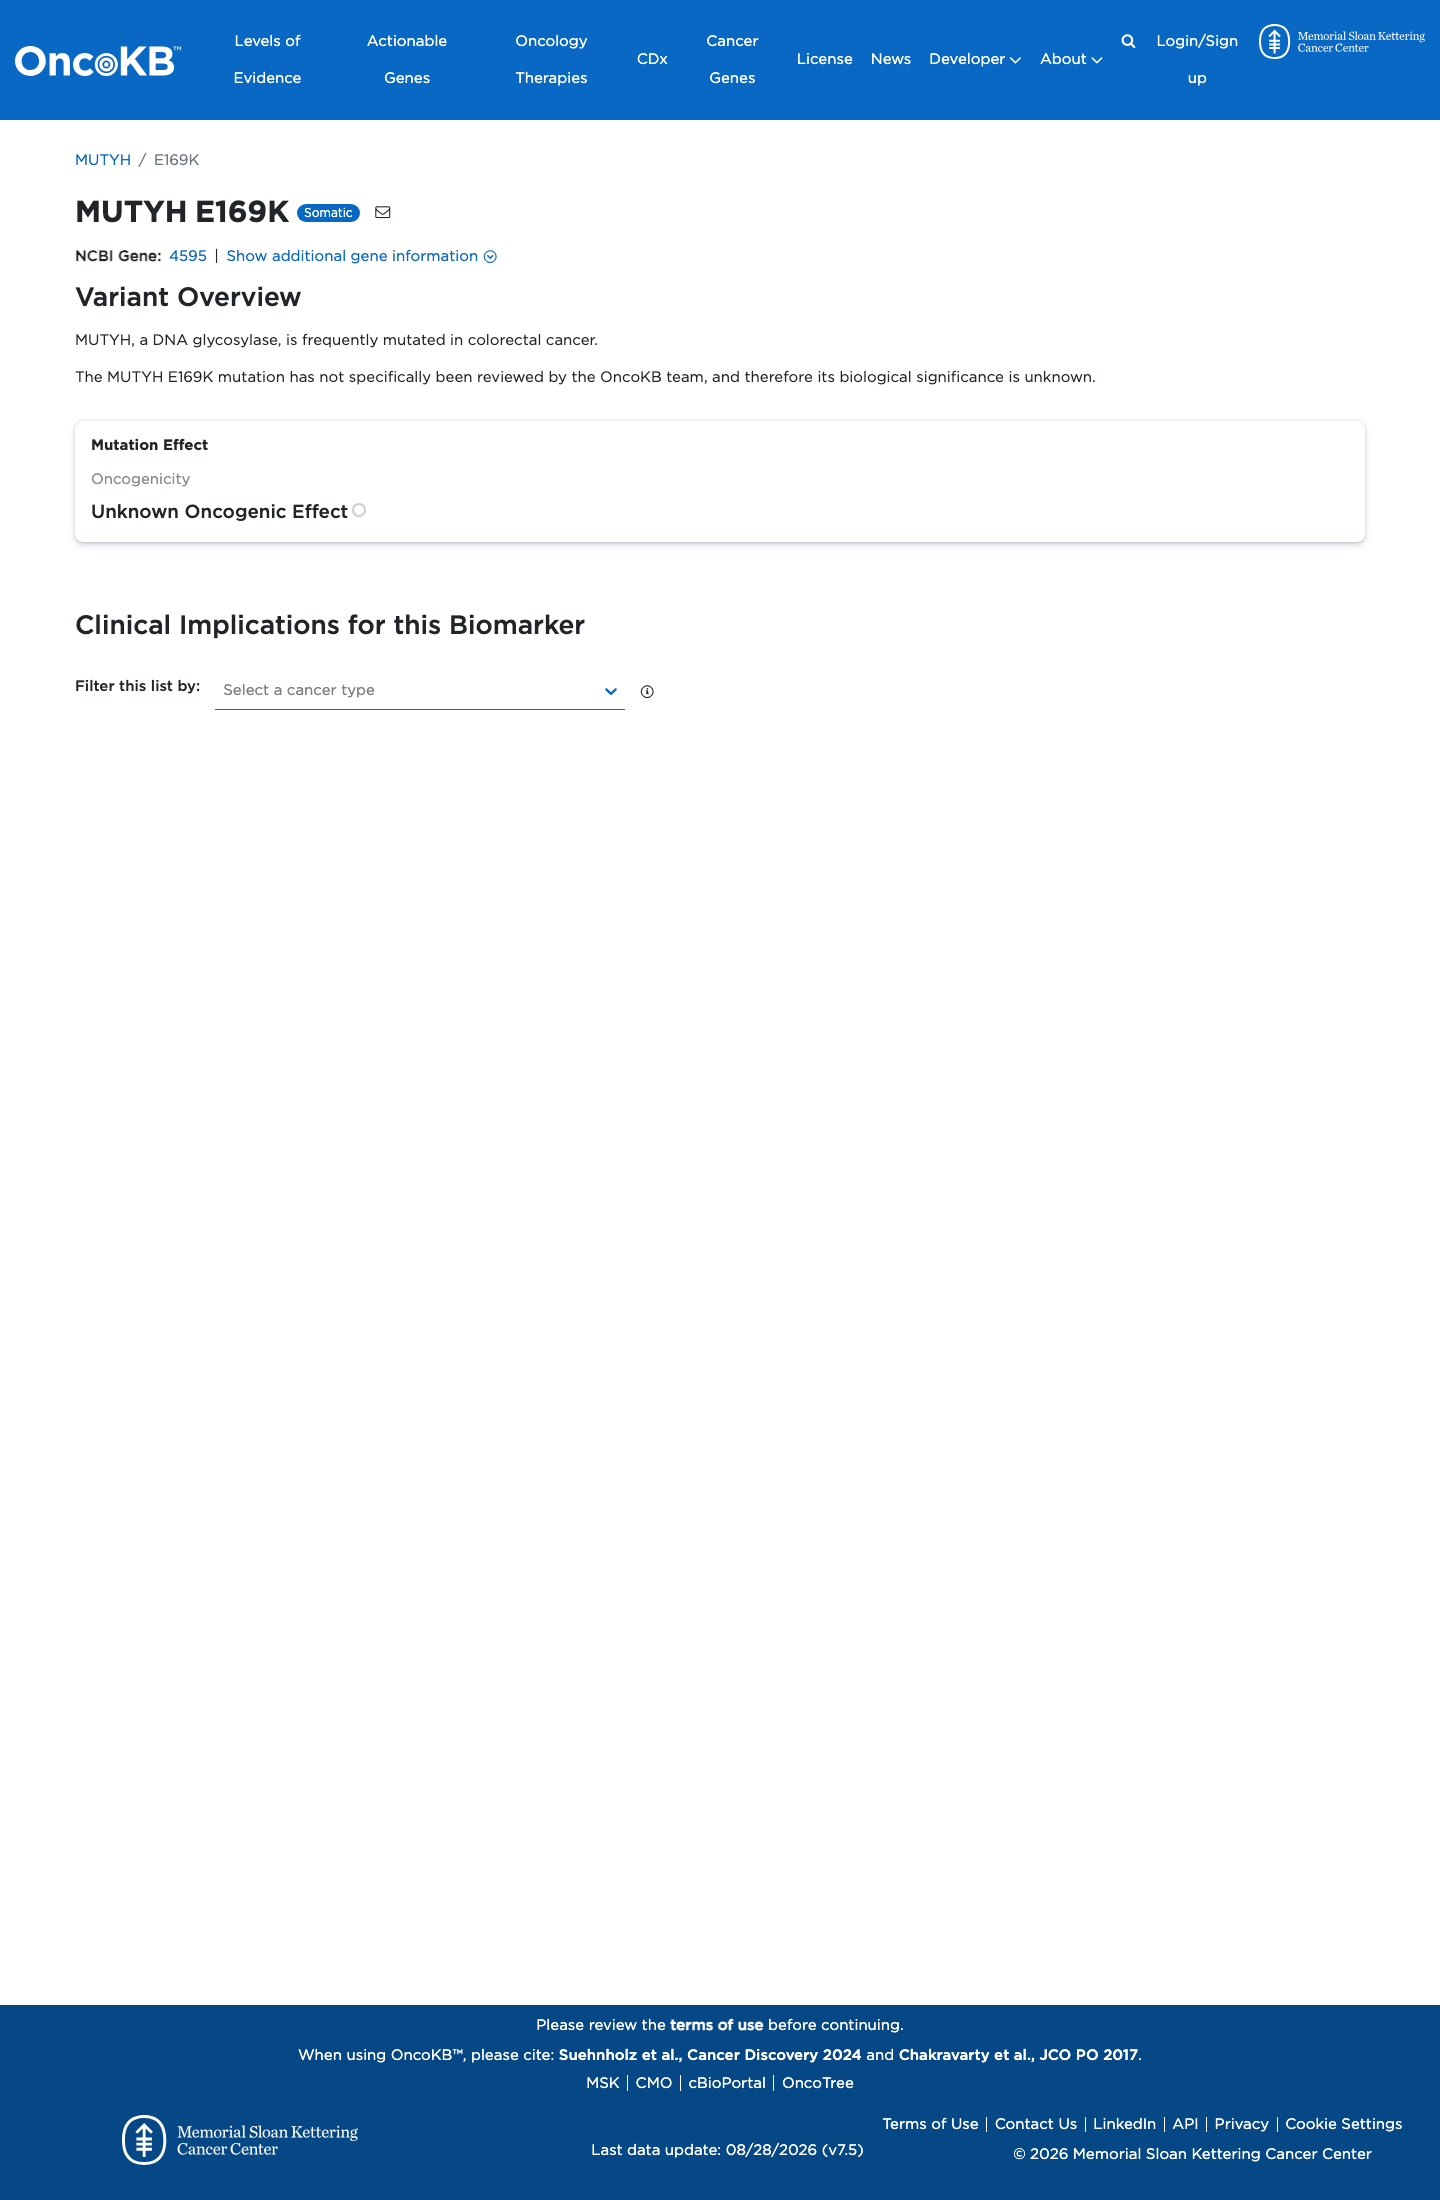
Functional / OncoKB screenshot
Functional Unknown Oncogenic Effect
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. MUTYH, a DNA glycosylase, is frequently mutated in colorectal cancer.
OncoKB ↗
COSMIC screenshot
COSMIC
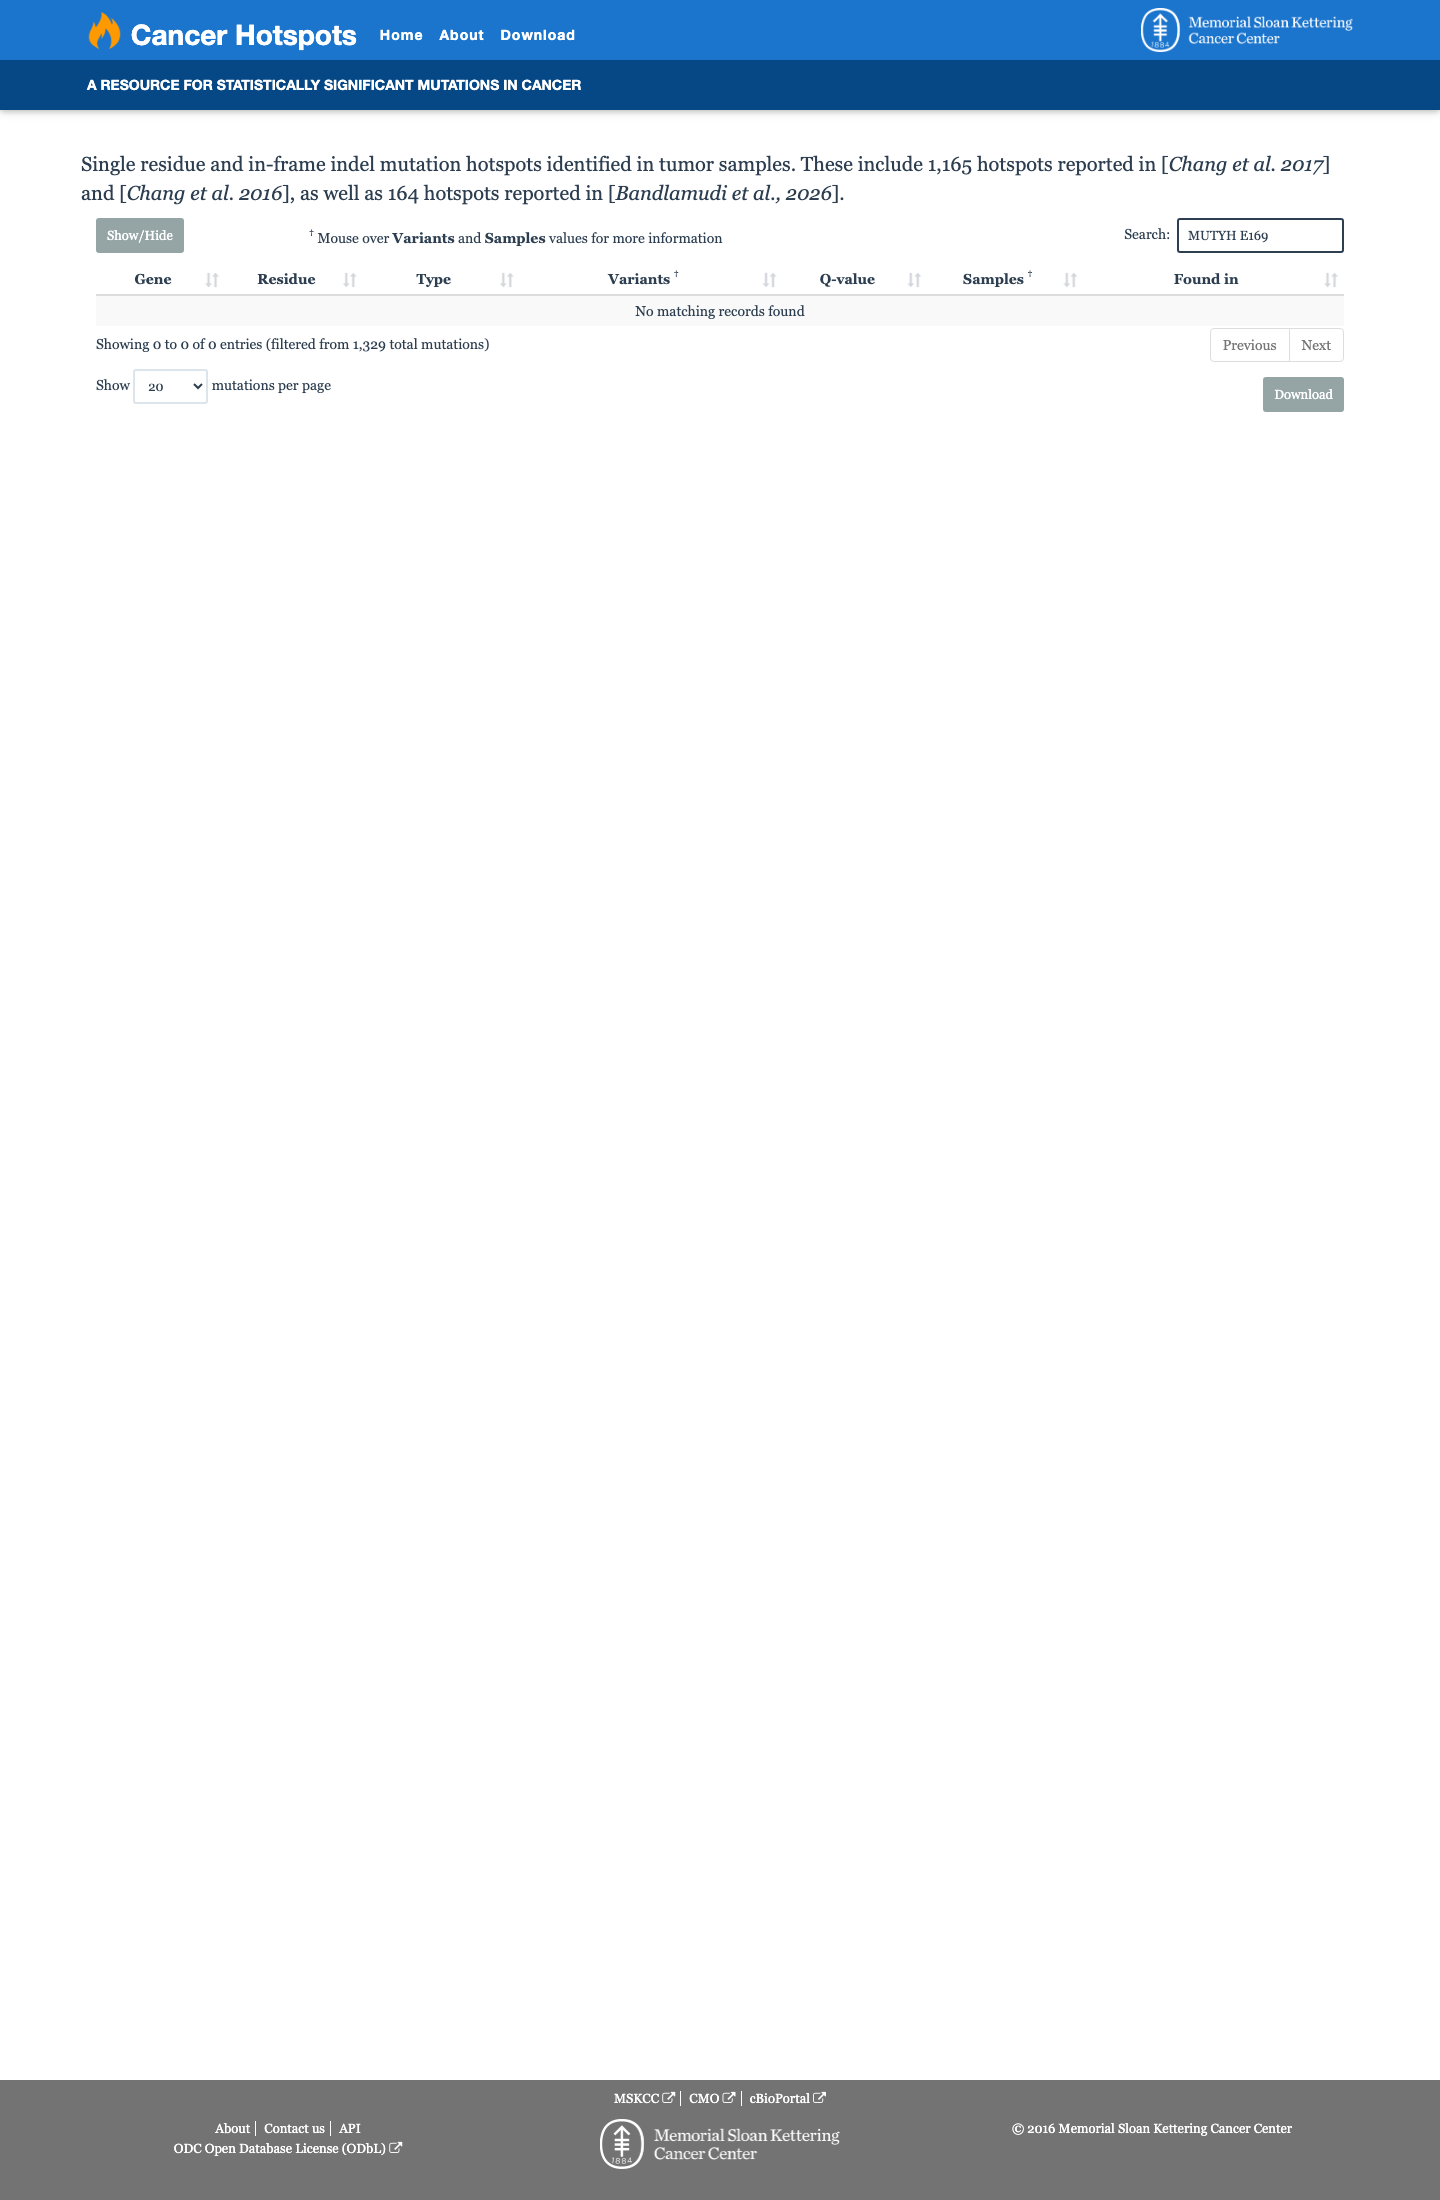
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Sources & reference links
9Sources
CSpec VCEP
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 5 PMIDs not cited in assessment
24310308 ↗ ACMG technical standards and guidelines for genetic testing for inherited colorectal cancer (Lynch syndrome, familial adenomatous polyposis, and MYH-associated polyposis). CLINVAR
25356965 ↗ ACMG policy statement: updated recommendations regarding analysis and reporting of secondary findings in clinical genome-scale sequencing. CLINVAR
25394175 ↗ A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment. CLINVAR
25452455 ↗ Hereditary colorectal cancer syndromes: American Society of Clinical Oncology Clinical Practice Guideline endorsement of the familial risk-colorectal cancer: European Society for Medical Oncology Clinical Practice Guidelines. CLINVAR
26389258 ↗ Cancer Genetics Risk Assessment and Counseling (PDQ®): Health Professional Versi CLINVAR