Back
NM_001128425.2:c.714C>T
p.Asn238= · MUTYH
0%
complete
Final classification
VUS
PM2
MUTYH
c.714C>T
p.Asn238=
synonymous · exon 9

MUTYH encodes a DNA repair enzyme (a glycosylase) that fixes oxidative DNA damage by removing adenine bases that have been mistakenly paired with guanine or with oxidized guanine lesions. Inherited mutations in both copies of the gene cause MUTYH-associated polyposis (MAP), a recessive condition that strongly predisposes people to multiple colorectal polyps and colorectal cancer. The gene acts as a tumor suppressor whose loss of function allows DNA damage to accumulate, and somatic changes in it have also been reported in colon cancer, though whether they drive cancer on their own is not fully established.

This variant

MUTYH encodes a DNA repair glycosylase, and biallelic loss-of-function variants cause the autosomal-recessive cancer-predisposition condition MUTYH-associated polyposis.

Transcript
NM_001128425.2
HGVS · transcript:coding
NM_001128425.2:c.714C>T
GRCh38
chr1:45332465 G>A
GRCh37
chr1:45798137 G>A
VUS: PM2 at supporting strength is the only applied criterion and does not satisfy a generic ACMG/AMP pathogenic, likely benign, or benign combination rule.
Classification rationale
PM2 VUS
MUTYH c.714C>T synonymous · exon 9

VUS: PM2 supporting is met because the gnomAD v4.1 allele frequency is 4.33708e-05, below the 0.0001 threshold, with zero homozygotes.

PM2 → VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_001128425.2 · variants mapped to exon structure
MUTYH NM_001128425.2
Fetching transcript structure from UCSC…
Applied criteria · 1 applied · 19 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 1
Strength Supporting Moderate Strong Very strong
PM2 supporting Pathogenic
Met at supporting strength: gnomAD v4.1 AF 4.33708e-05 is below the PM2 threshold of 0.0001, with zero homozygotes.
gnomAD v2.1 non-cancer exomes: 2/236,434 alleles, AF 8.45902e-06, homozygotes 0.gnomAD v3.1 non-cancer genomes: 8/147,954 alleles, AF 5.40709e-05, homozygotes 0.gnomAD v4.1 combined exome-plus-genome data: 70/1,613,988 alleles, AF 4.33708e-05, homozygotes 0.
Assessed · not applied · 8 not met · 11 not assessed
Pathogenic
PS2 Not assessed: no proband disease, family-history, or confirmed maternity-and-paternity evidence is available for this variant.
PS3 Not assessed: no variant-specific validated functional assay was identified for synonymous MUTYH c.714C>T (p.Asn238=).
PS4 Not assessed: no variant-specific case-control odds ratio or relative risk, and no confidence interval, is available to evaluate the >5.0 and non-inclusion-of-1.0 thresholds.
PM3 Not assessed: no documented parent/offspring phasing or affected-proband observation places c.714C>T in trans with a pathogenic variant.
PM6 Not assessed: no assumed de novo observation or parental testing is documented for the evaluated variant.
PP1 Not assessed: no affected relatives or informative segregation data are documented for the evaluated variant.
PP3 Not met: synonymous-variant SpliceAI max delta 0.114 is below the PP3 supporting threshold of >= 0.2.
PP4 Not assessed: no patient phenotype or family history is provided to establish a highly specific presentation for a single-gene disease.
PP5 Not met: ClinVar has 0 expert-panel submissions for the exact variant, and available laboratory assertions are not eligible to trigger PP5.
Benign
BA1 Not met: the highest observed allele frequency is 0.00049456, far below the BA1 stand-alone threshold of 0.05.
BS1 Not met: the highest observed allele frequency is 0.00049456, below the generic BS1 threshold of 0.01.
BS2 Not met: zero homozygotes are reported across the available gnomAD datasets, so no qualifying healthy adult homozygote is observed.
BS3 Not assessed: no variant-specific validated functional assay demonstrated no damaging effect for synonymous MUTYH c.714C>T (p.Asn238=).
BS4 Not assessed: no tested affected relatives are documented, so lack of segregation cannot be evaluated.
BP2 Not assessed: no documented cis or trans co-occurrence with a pathogenic variant is available for c.714C>T.
BP4 Not met: synonymous-variant SpliceAI max delta 0.114 exceeds the BP4 supporting threshold of <= 0.1.
BP5 Not assessed: no case-level evidence links this variant to a patient with an independently established alternate molecular basis for disease.
BP6 Not met: ClinVar has 0 expert-panel submissions, so its ordinary laboratory Likely benign and Benign assertions cannot trigger BP6.
BP7 Not met: synonymous variant SpliceAI max delta 0.114 exceeds the <= 0.1 no-impact cutoff required for BP7.
N/A · 8 PVS1 · PS1 · PM1 · PM4 · PM5 · PP2 · BP1 · BP3
Research & evidence
Population frequency
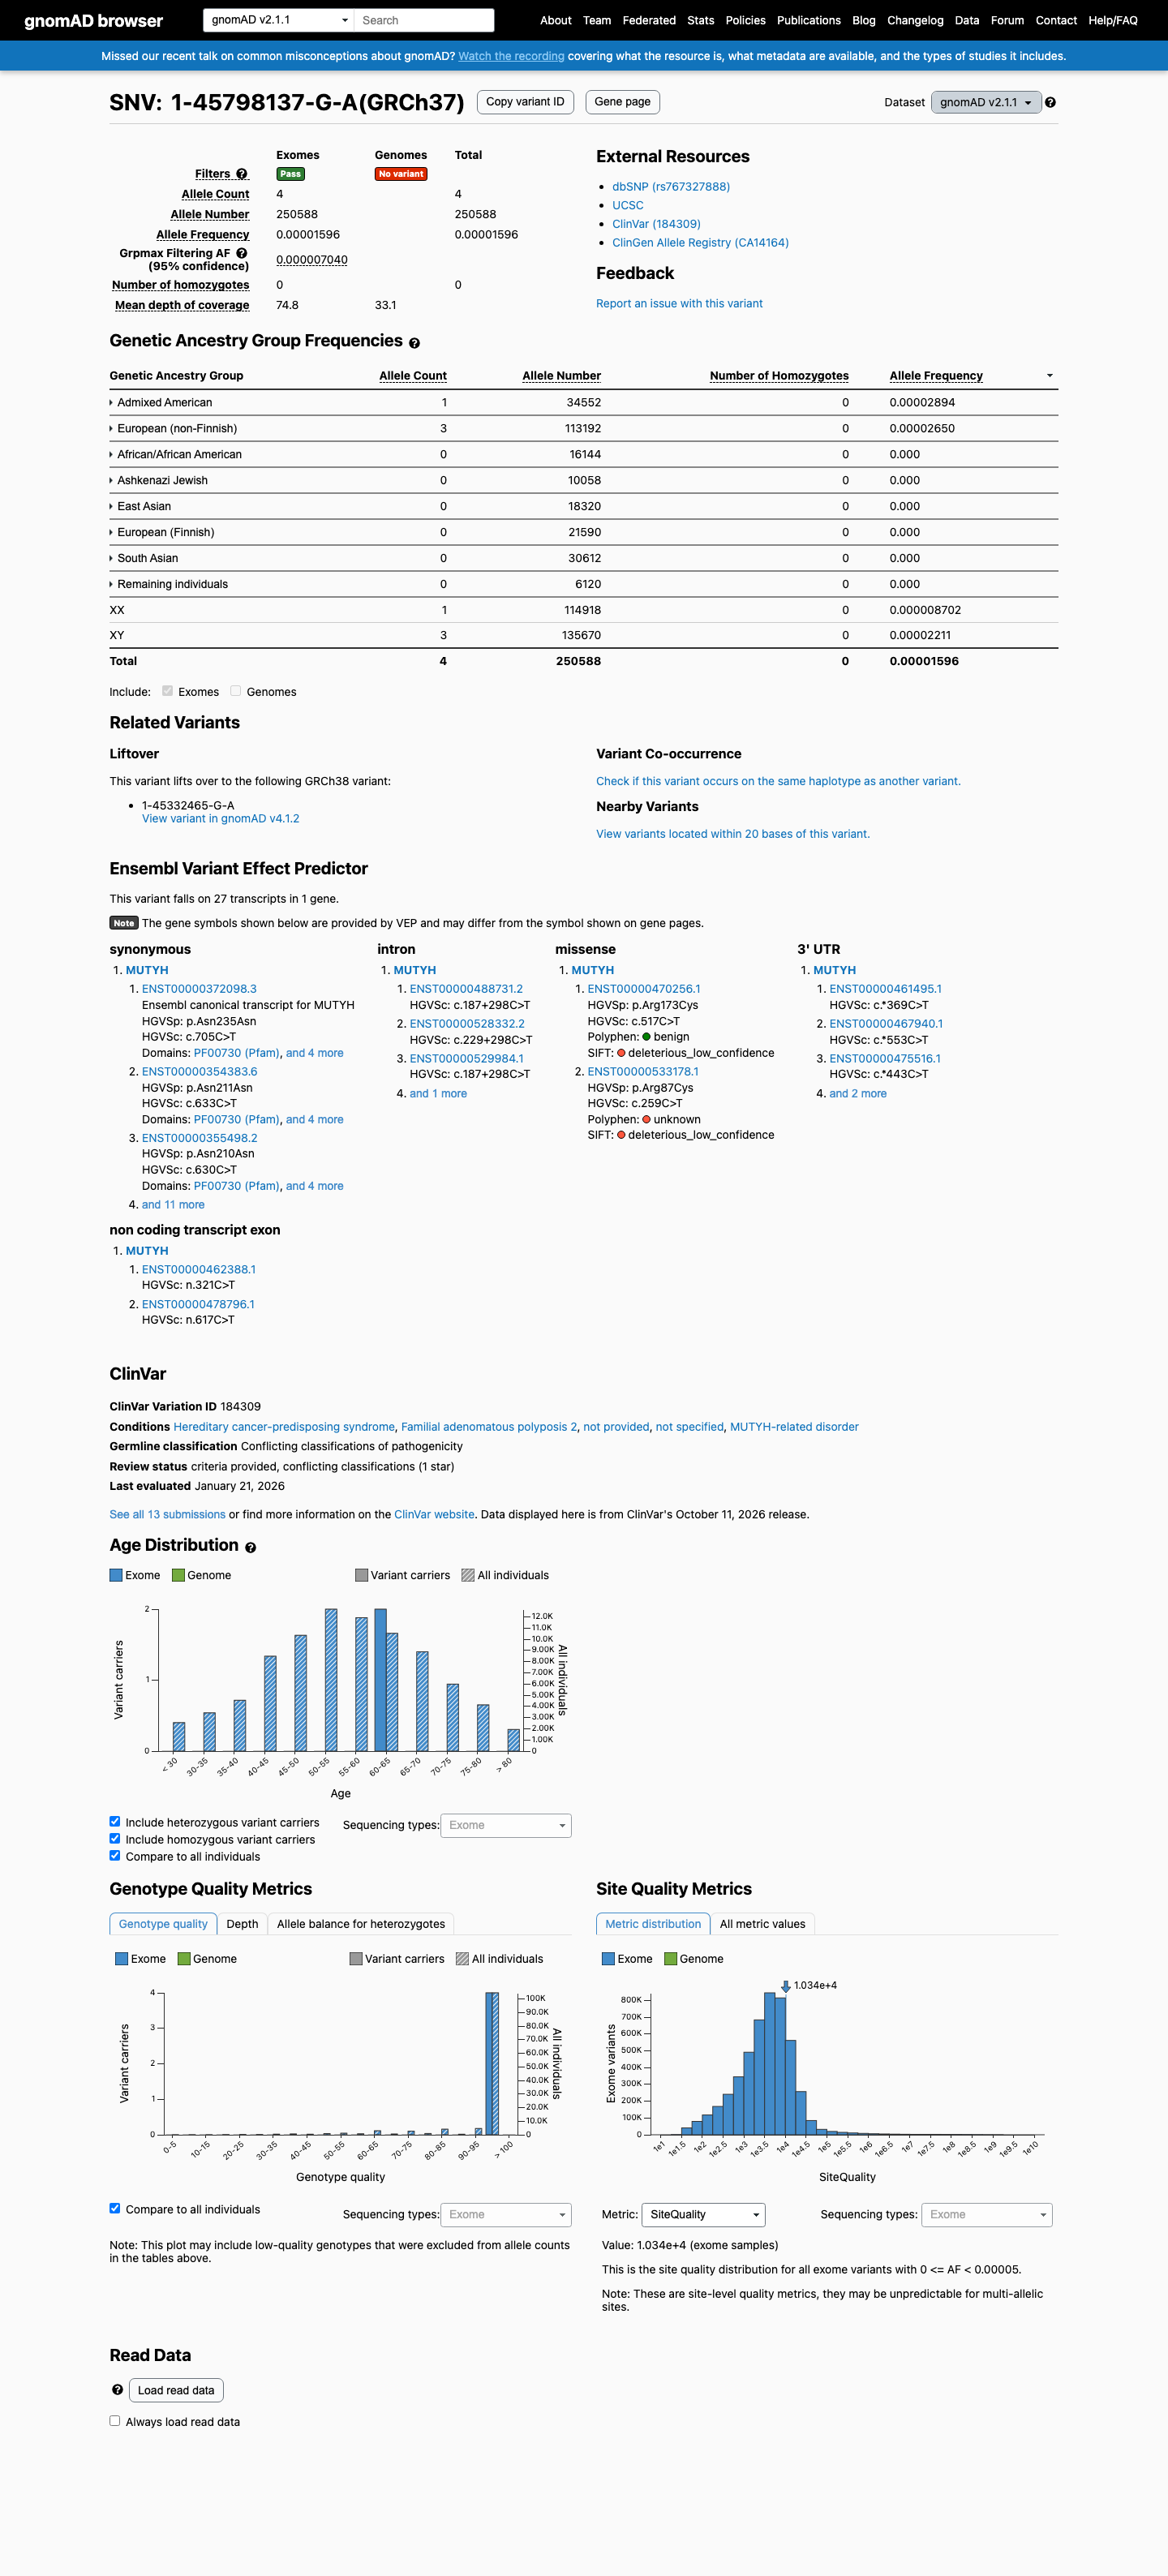
gnomAD v4.1 screenshot
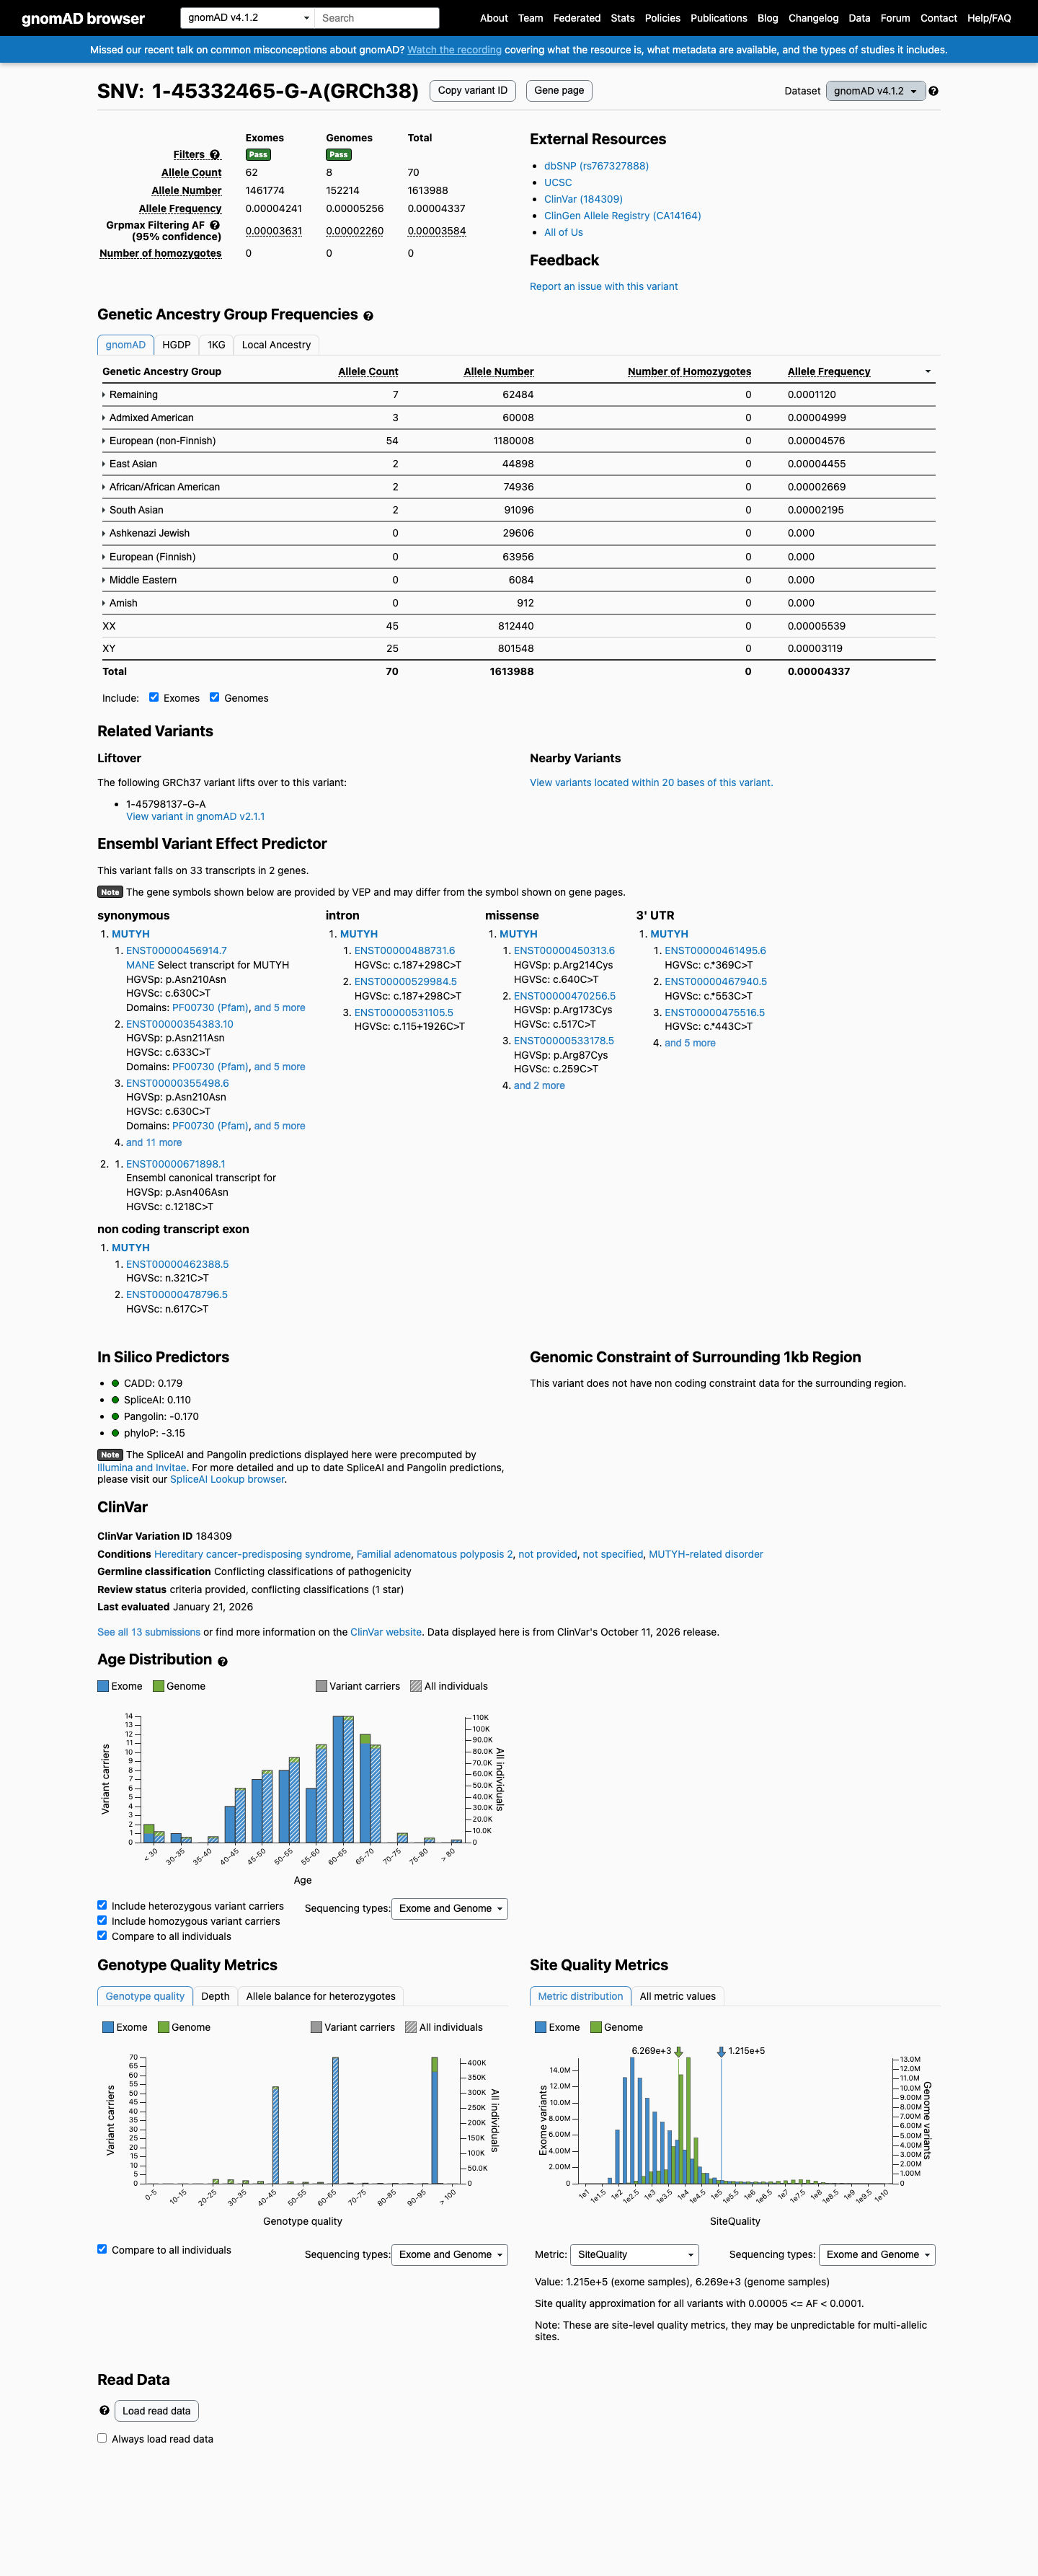
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 4.33708e-05; MAF= 0.00434%, 70/1613988 alleles, homozygotes = 0) and has highest observed frequency in the Remaining individuals population (AF= 0.000112029; MAF= 0.01120%, 7/62484 alleles, homozygotes = 0); grpmax FAF= 3.584e-05.
v2.1
This variant is present in gnomAD v2.1 (AF= 1.59625e-05; MAF= 0.00160%, 4/250588 alleles, homozygotes = 0) and has highest observed frequency in the Admixed American population (AF= 2.89419e-05; MAF= 0.00289%, 1/34552 alleles, homozygotes = 0); grpmax FAF= 7.04e-06.
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.0043% · 70 / 1,613,988
0 hom · FAF 0.0036%
Remaining individuals
7 / 62,484
0.011%
Admixed American
3 / 60,008
0.005%
European (non-Finnish)
54 / 1,180,008
0.0046%
East Asian
2 / 44,898
0.0045%
African/African American
2 / 74,936
0.0027%
South Asian
2 / 91,096
0.0022%
+ 4 not observed (European (Finnish), Amish, Middle Eastern, Ashkenazi Jewish)
gnomAD v2.1
0.0016% · 4 / 250,588
0 hom · FAF 0.0007%
Admixed American
1 / 34,552
0.0029%
European (non-Finnish)
3 / 113,192
0.0027%
+ 6 not observed (African/African American, Ashkenazi Jewish, East Asian, European (Finnish), Remaining individuals, South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
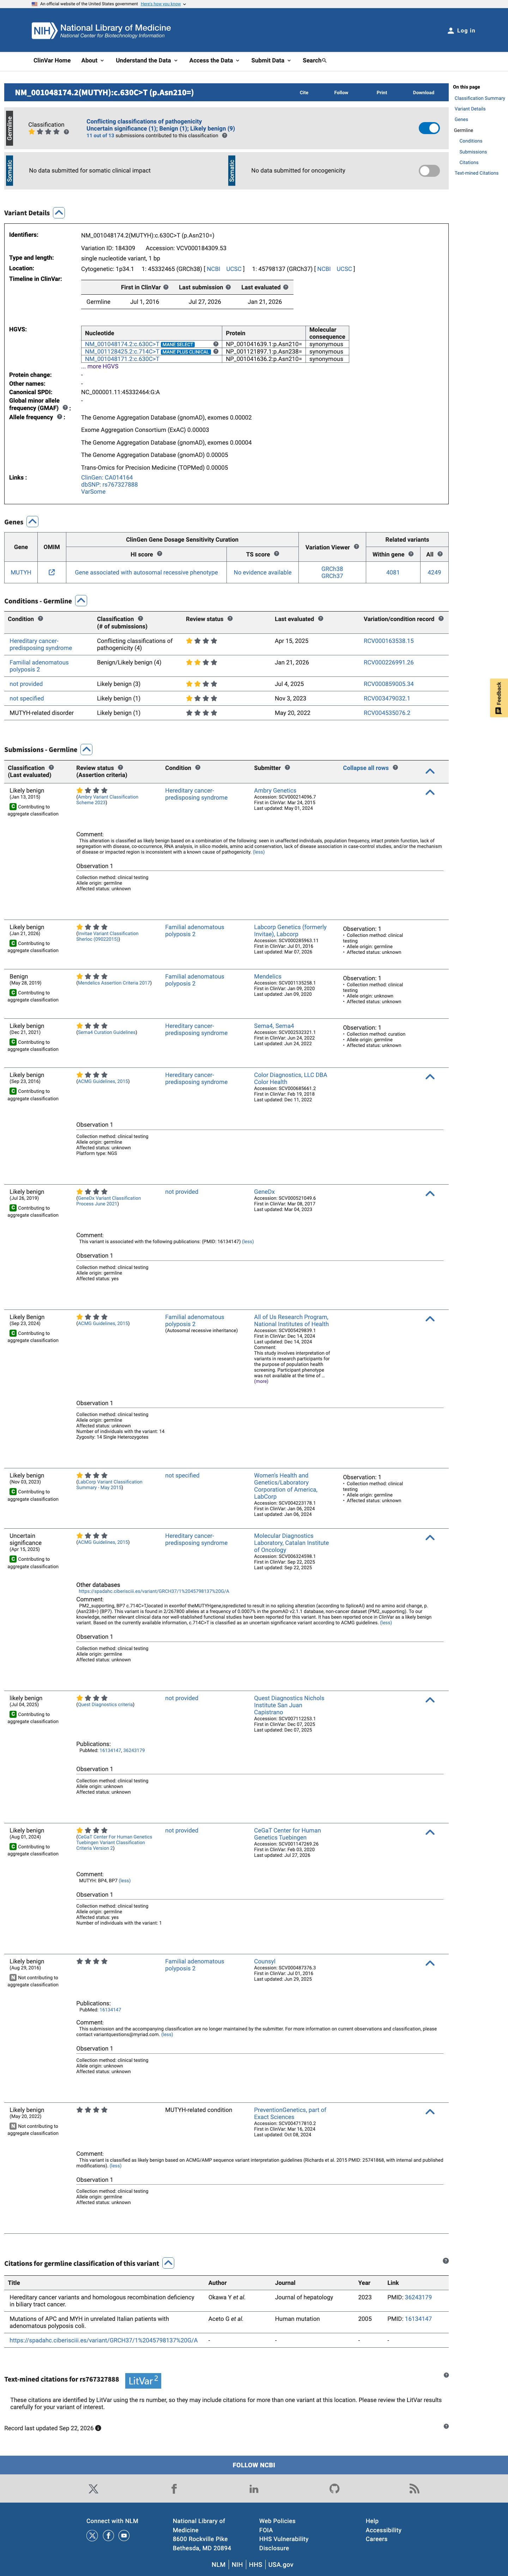
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Likely benign (8 clinical laboratories) and as Benign (1 clinical laboratory) and as Likely Benign (1 clinical laboratory) and as Uncertain significance (1 clinical laboratory) and as likely benign (1 clinical laboratory). (ClinVarID = 184309)
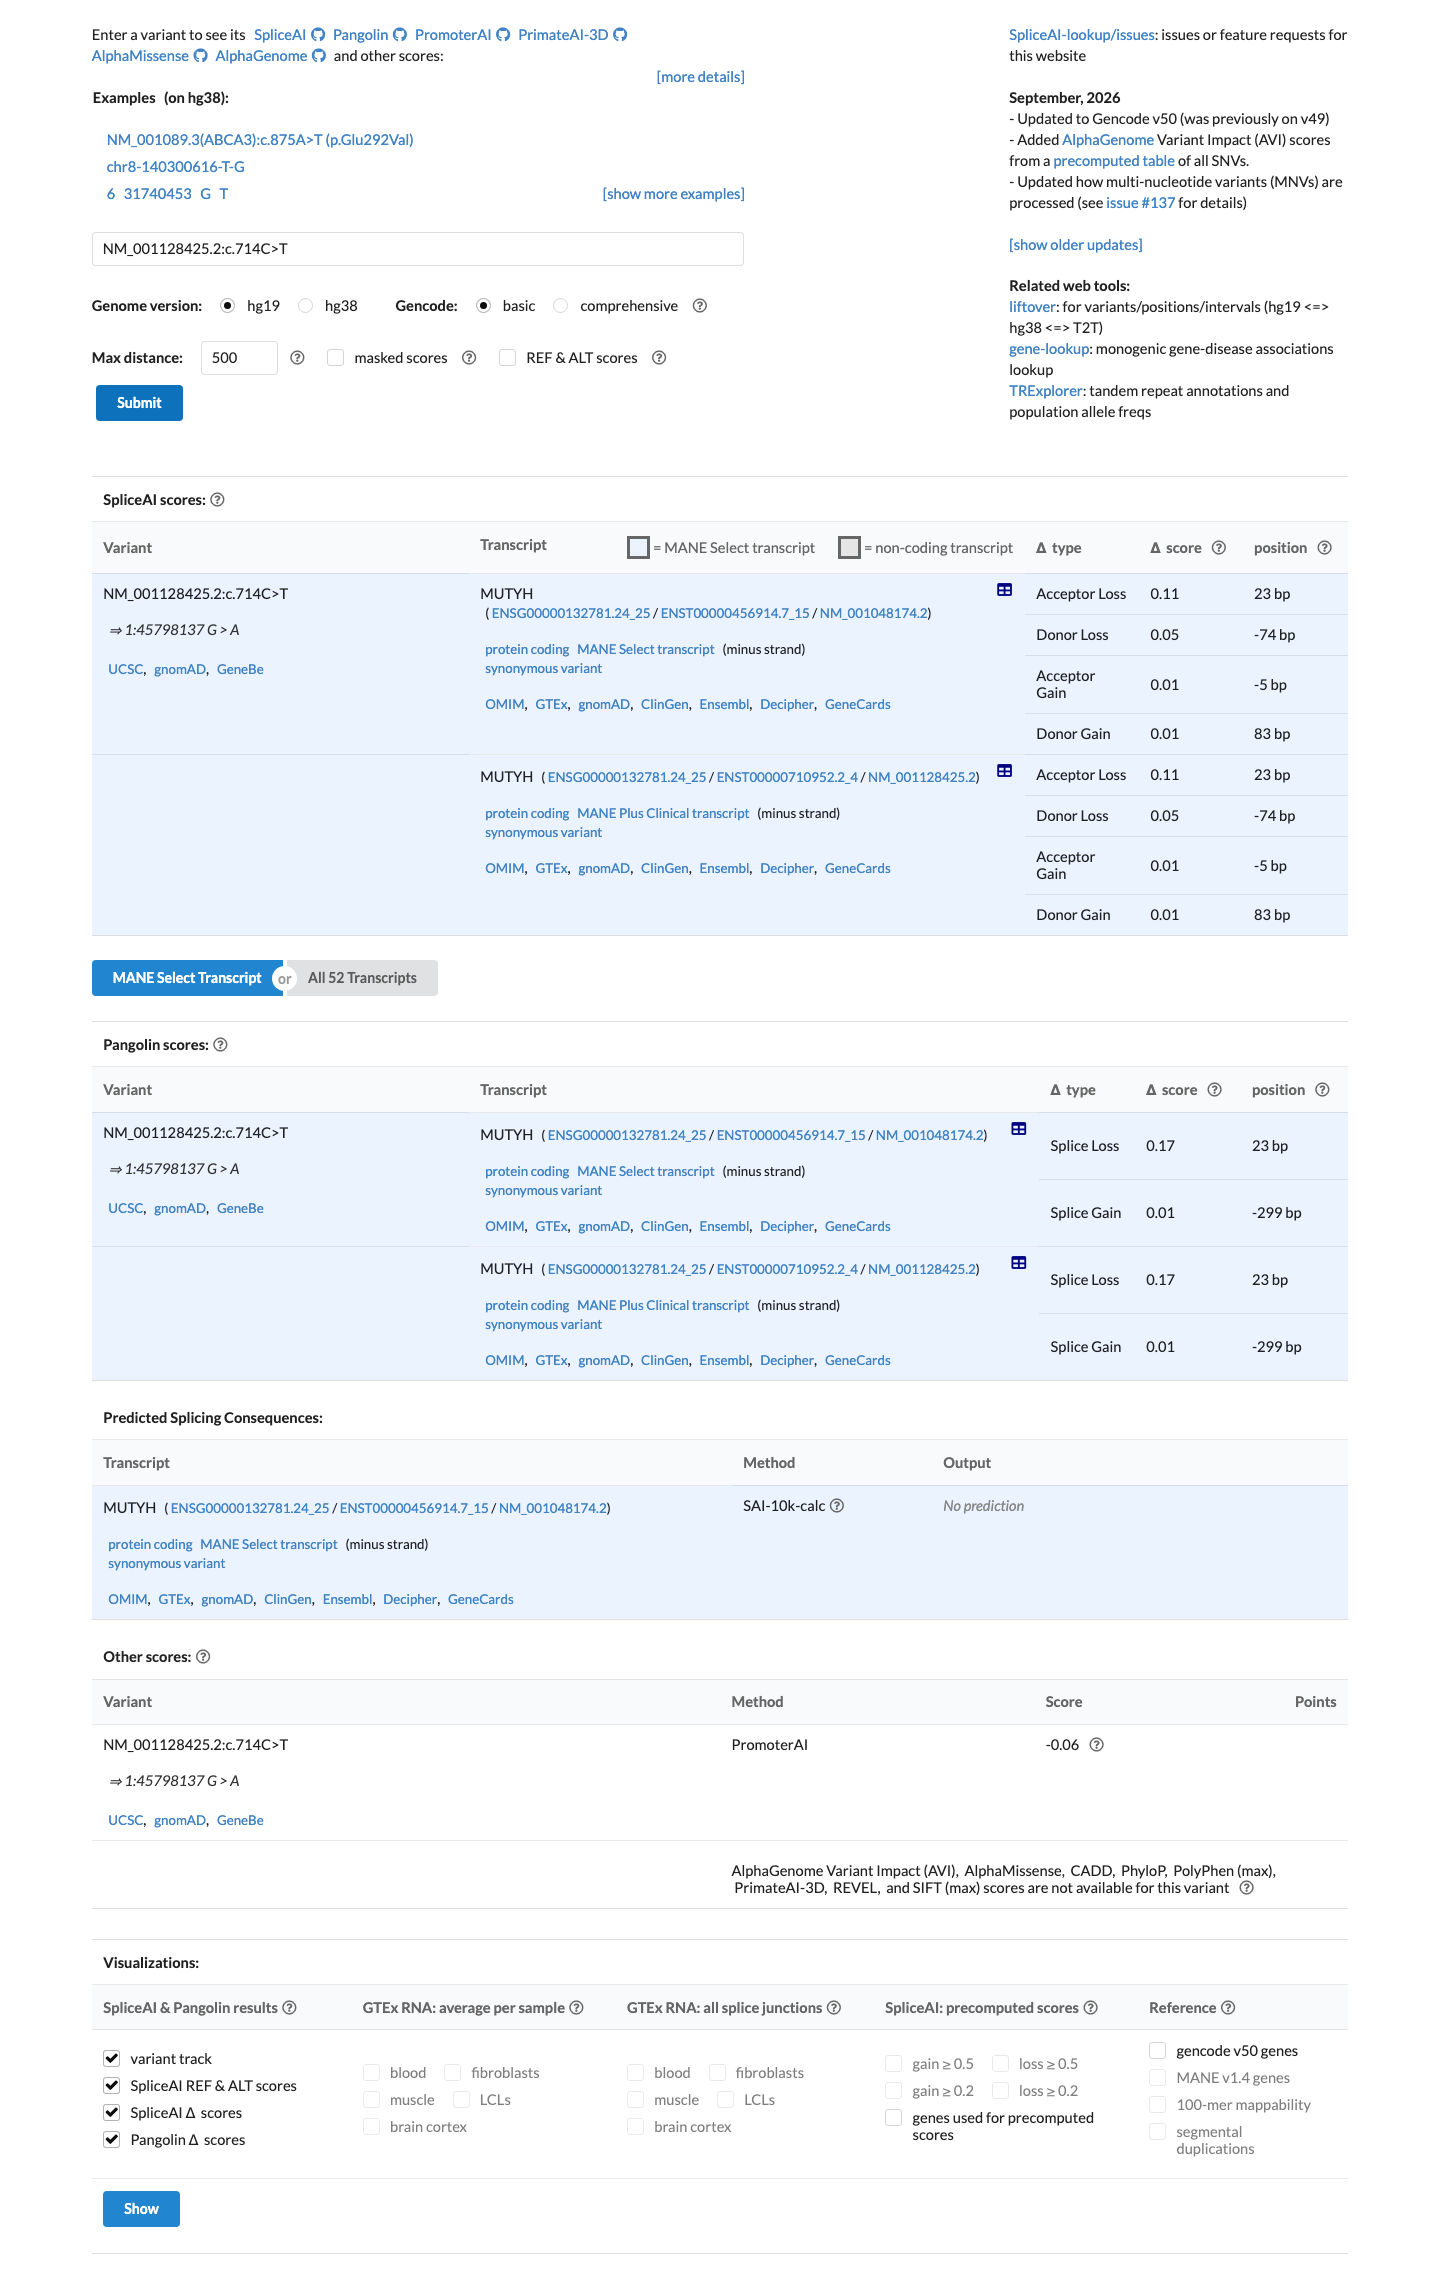
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.11).
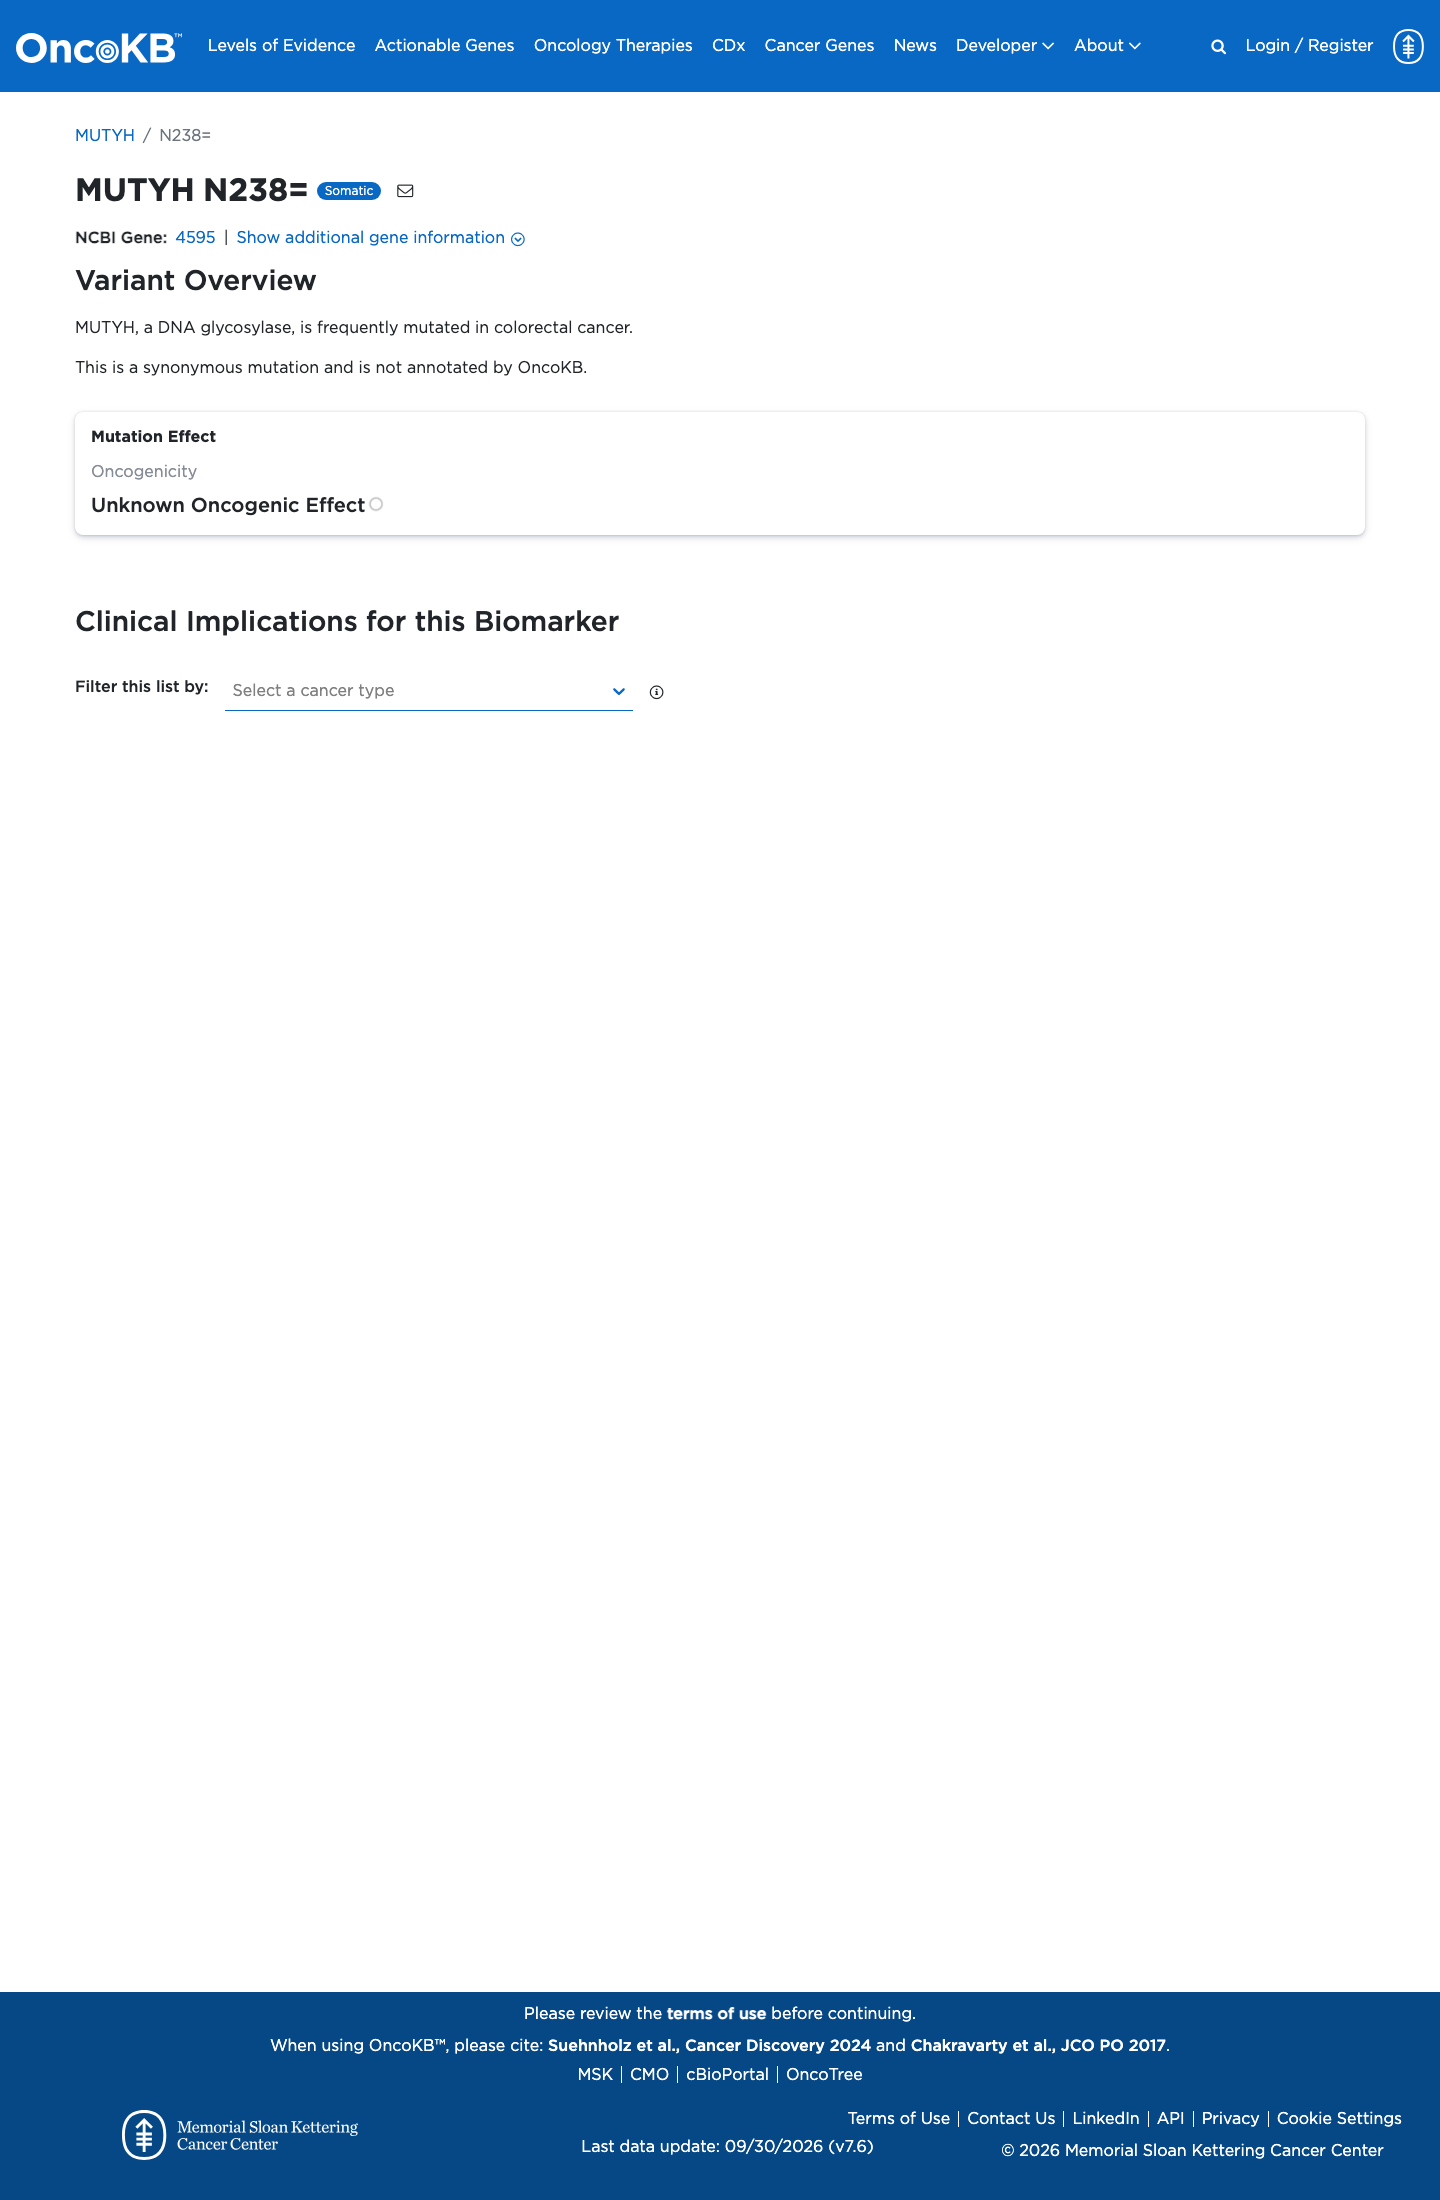
Functional / OncoKB screenshot
Functional Unknown Oncogenic Effect
OncoKB identified curated literature and non-variant-specific oncogenicity context for review; listed oncogenicity label: Unknown Oncogenic Effect.
OncoKB ↗
COSMIC screenshot
COSMIC
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
1papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 8 further PMIDs triaged but not cited — see Sources & references.
Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.
Found
ACMG/AMP 2015 criterion definition PMID:25741868).
Applied to
→PM2 supporting
Sources & reference links
9Sources
CSpec VCEP
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 8 PMIDs not cited in assessment
16134147 ↗ Mutations of APC and MYH in unrelated Italian patients with adenomatous polyposis coli. CLINVAR
26467025 ↗ A Standardized DNA Variant Scoring System for Pathogenicity Assessments in Mendelian Disorders. CLINVAR
23035301 ↗ MUTYH Polyposis. CLINVAR
23788249 ↗ ACMG recommendations for reporting of incidental findings in clinical exome and genome sequencing. CLINVAR
25356965 ↗ ACMG policy statement: updated recommendations regarding analysis and reporting of secondary findings in clinical genome-scale sequencing. CLINVAR
25394175 ↗ A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment. CLINVAR
27854360 ↗ Recommendations for reporting of secondary findings in clinical exome and genome sequencing, 2016 update (ACMG SF v2.0): a policy statement of the American College of Medical Genetics and Genomics. CLINVAR
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification criteria. CLINVAR