Back
NM_002439.5:c.1360C>T
p.Arg454Ter · MSH3
ACMG/AMP
0%
complete
Final classification
Likely Pathogenic
PVS1PM2
MSH3
c.1360C>T
p.Arg454Ter
nonsense · exon 9

MSH3 encodes a protein that partners with MSH2 to form MutS beta, a key component of the DNA mismatch repair system that recognizes and corrects errors made during DNA replication. It acts as a tumor suppressor: loss of MSH3 function leads to mismatch repair deficiency, an elevated mutation rate, and increased cancer risk, and the gene is frequently altered in mismatch repair-deficient colorectal cancers. Defects in MSH3 are linked to susceptibility to endometrial cancer, and inherited mutations are associated with an autosomal recessive form of familial adenomatous polyposis, with minimal evidence for a role in Lynch syndrome. Tumors with mismatch repair deficiency, including those involving MSH3, tend to respond well to immune checkpoint inhibitor therapies.

This variant

MSH3 encodes the MutS beta mismatch-repair partner of MSH2, so this truncating p.Arg454Ter allele is expected to impair mismatch-repair function, the loss of which underlies the autosomal-recessive colorectal adenomatous polyposis (familial adenomatous polyposis 4) and mismatch-repair-deficient cancer predisposition associated with the gene.
Transcript
NM_002439.5
HGVS · transcript:coding
NM_002439.5:c.1360C>T
GRCh38
chr5:80725472 C>T
GRCh37
chr5:80021291 C>T
Likely Pathogenic: PVS1 (very strong, NMD-predicted p.Arg454Ter truncating ~60% of MSH3) plus PM2 (supporting) meet the generic ACMG/AMP 2015 one-very-strong-plus-supporting rule.
Classification rationale
PVS1PM2 Likely Pathogenic
MSH3 c.1360C>T nonsense · exon 9

Likely Pathogenic: PVS1 very strong is met because p.Arg454Ter is an NMD-predicted exon 9 of 24 nonsense removing about 60% of MSH3, a gene where loss of function causes disease. Likely Pathogenic: PM2 supporting is met because gnomAD v2.1 (2.79e-05) and v4.1 (1.24e-05) frequencies are below the 0.0001 cutoff with zero homozygotes. PS3 and PS4 not met: no functional assay and no case-control enrichment exist for this allele. PP5 not met: ClinVar variation 644460 has eight laboratory submissions but zero expert-panel submissions. BA1, BS1, BS2 and BP6 not met: the highest population frequency (about 0.13% in a Korean sub-cohort) is far below the 5% and 1% thresholds, no homozygote is observed, and no expert panel has classified the variant. Final combination: PVS1 (very strong) plus one supporting criterion (PM2) reaches the Likely Pathogenic rule, while PVS1 alone would not have reached Pathogenic.

PVS1 + PM2 Likely Pathogenic
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_002439.5 · variants mapped to exon structure
MSH3 NM_002439.5
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 16 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
PVS1 very strong Pathogenic
Met at very strong: nonsense stop at codon 454 of 1137 in exon 9 of 24, NMD-predicted, deleting about 60% of MSH3 including its DNA-recognition and dimerization domains.
Normalization/mutalyzer: NM_002439.5:c.1360C>T -> NM_002439.5(NP_002430.3):p.(Arg454Ter); protein first position 453/454, last predicted position 454 versus last original position 1138; the reference MSH3 protein is 1137 residues with CDS c.1-3414.VariantValidator: NM_002439.5 is the MANE Select and RefSeq Select transcript for MSH3 (HGNC:7326, OMIM 600887, 5q14.1); the variant maps to exon 9 of 24 in NG_016607.2, NC_000005.9 and NC_000005.10.Transcript selector exon boundaries: exon 9 spans c.1341-1453, so c.1360 sits 20 nt into exon 9; the penultimate exon 23 spans c.3131-3302 and the terminal exon 24 spans c.3303-*953, placing the premature stop roughly 1.9 kb 5' of the last 50 bp of the penultimate exon, hence NMD is predicted.
PM2 supporting review Pathogenic
Met at Supporting: gnomAD v2.1 and v4.1 frequencies (0.0000279, 0.0000124) sit below the 0.0001 threshold with zero homozygotes.
PM2 Supporting threshold applied: AF <= 0.0001 (ClinGen SVI 2020 recommendation to downgrade PM2 to Supporting; PMID:25741868).gnomAD v2.1 all-comers: AF 2.78689e-05 (7/251,176 alleles), 0 homozygotes; highest ancestry East Asian AF 0.000272183 (5/18,370); grpmax FAF 0.00010636.gnomAD v4.1 all-comers: AF 1.23992e-05 (20/1,613,002 alleles), 0 homozygotes; highest ancestry East Asian AF 0.000133917 (6/44,804); grpmax FAF 5.796e-05.
Assessed · not applied · 8 not met · 8 not assessed
Pathogenic
PS2 Not assessed: no parental or trio testing exists in the case evidence, so de novo status of c.1360C>T cannot be established.
PS3 Not met: no functional assay of p.Arg454Ter exists - the case's MSH3 functional data (HEK293T/western blot, EMAST) characterise other loss-of-function alleles, not codon 454.
PS4 Not met: no case-control enrichment data exist for this variant, and no odds ratio is reported (gnomAD v4.1 AF 1.24e-05, 20/1,613,002 alleles).
PM3 Not assessed: no proband genotype or phase data exist, so c.1360C>T cannot be shown in trans with a pathogenic MSH3 allele.
PM6 Not assessed: no parental genotypes are available, so c.1360C>T cannot be assumed de novo at any strength.
PP1 Not assessed: no pedigree, affected relatives or family genotypes are reported, so co-segregation of c.1360C>T cannot be evaluated.
PP4 Not assessed: no proband phenotype, HPO terms, or family history was provided, so phenotype specificity for MSH3-associated polyposis cannot be evaluated.
PP5 Not met: ClinVar variation 644460 has zero expert-panel submissions, all 8 being single-submitter laboratories, so the exact-variant expert-panel requirement is unmet.
Benign
BA1 Not met: highest gnomAD allele frequency of 0.13 percent (East Asian Korean subset) remains far below the 0.05 BA1 stand-alone threshold.
BS1 Not met: gnomAD v2.1 allele frequency 0.0000279 is roughly 360-fold below the 0.01 BS1 strong threshold.
BS2 Not met: zero homozygotes in gnomAD (v2.1 7/251,176; v4.1 20/1,613,002) means no healthy homozygous adult supports this recessive variant.
BS3 Not met: no functional assay reports normal MSH3 function for p.Arg454Ter - the case's MSH3 assays cover other alleles and show loss, not preservation, of function.
BS4 Not assessed: no family segregation data exist, so non-segregation of c.1360C>T in affected relatives cannot be documented.
BP2 Not assessed: no phase data exist in any source, so c.1360C>T cannot be shown in cis with a pathogenic variant.
BP5 Not assessed: no proband-level data on co-occurring variants or phenotype was provided, so an alternate molecular basis for disease cannot be evaluated.
BP6 Not met: ClinVar variation 644460 has zero expert-panel submissions, and all 8 laboratory submissions classify the variant Pathogenic or Likely pathogenic.
N/A · 10 PS1 · PM1 · PM4 · PM5 · PP2 · PP3 · BP1 · BP3 · BP4 · BP7
Research & evidence
Population frequency · supports pathogenic
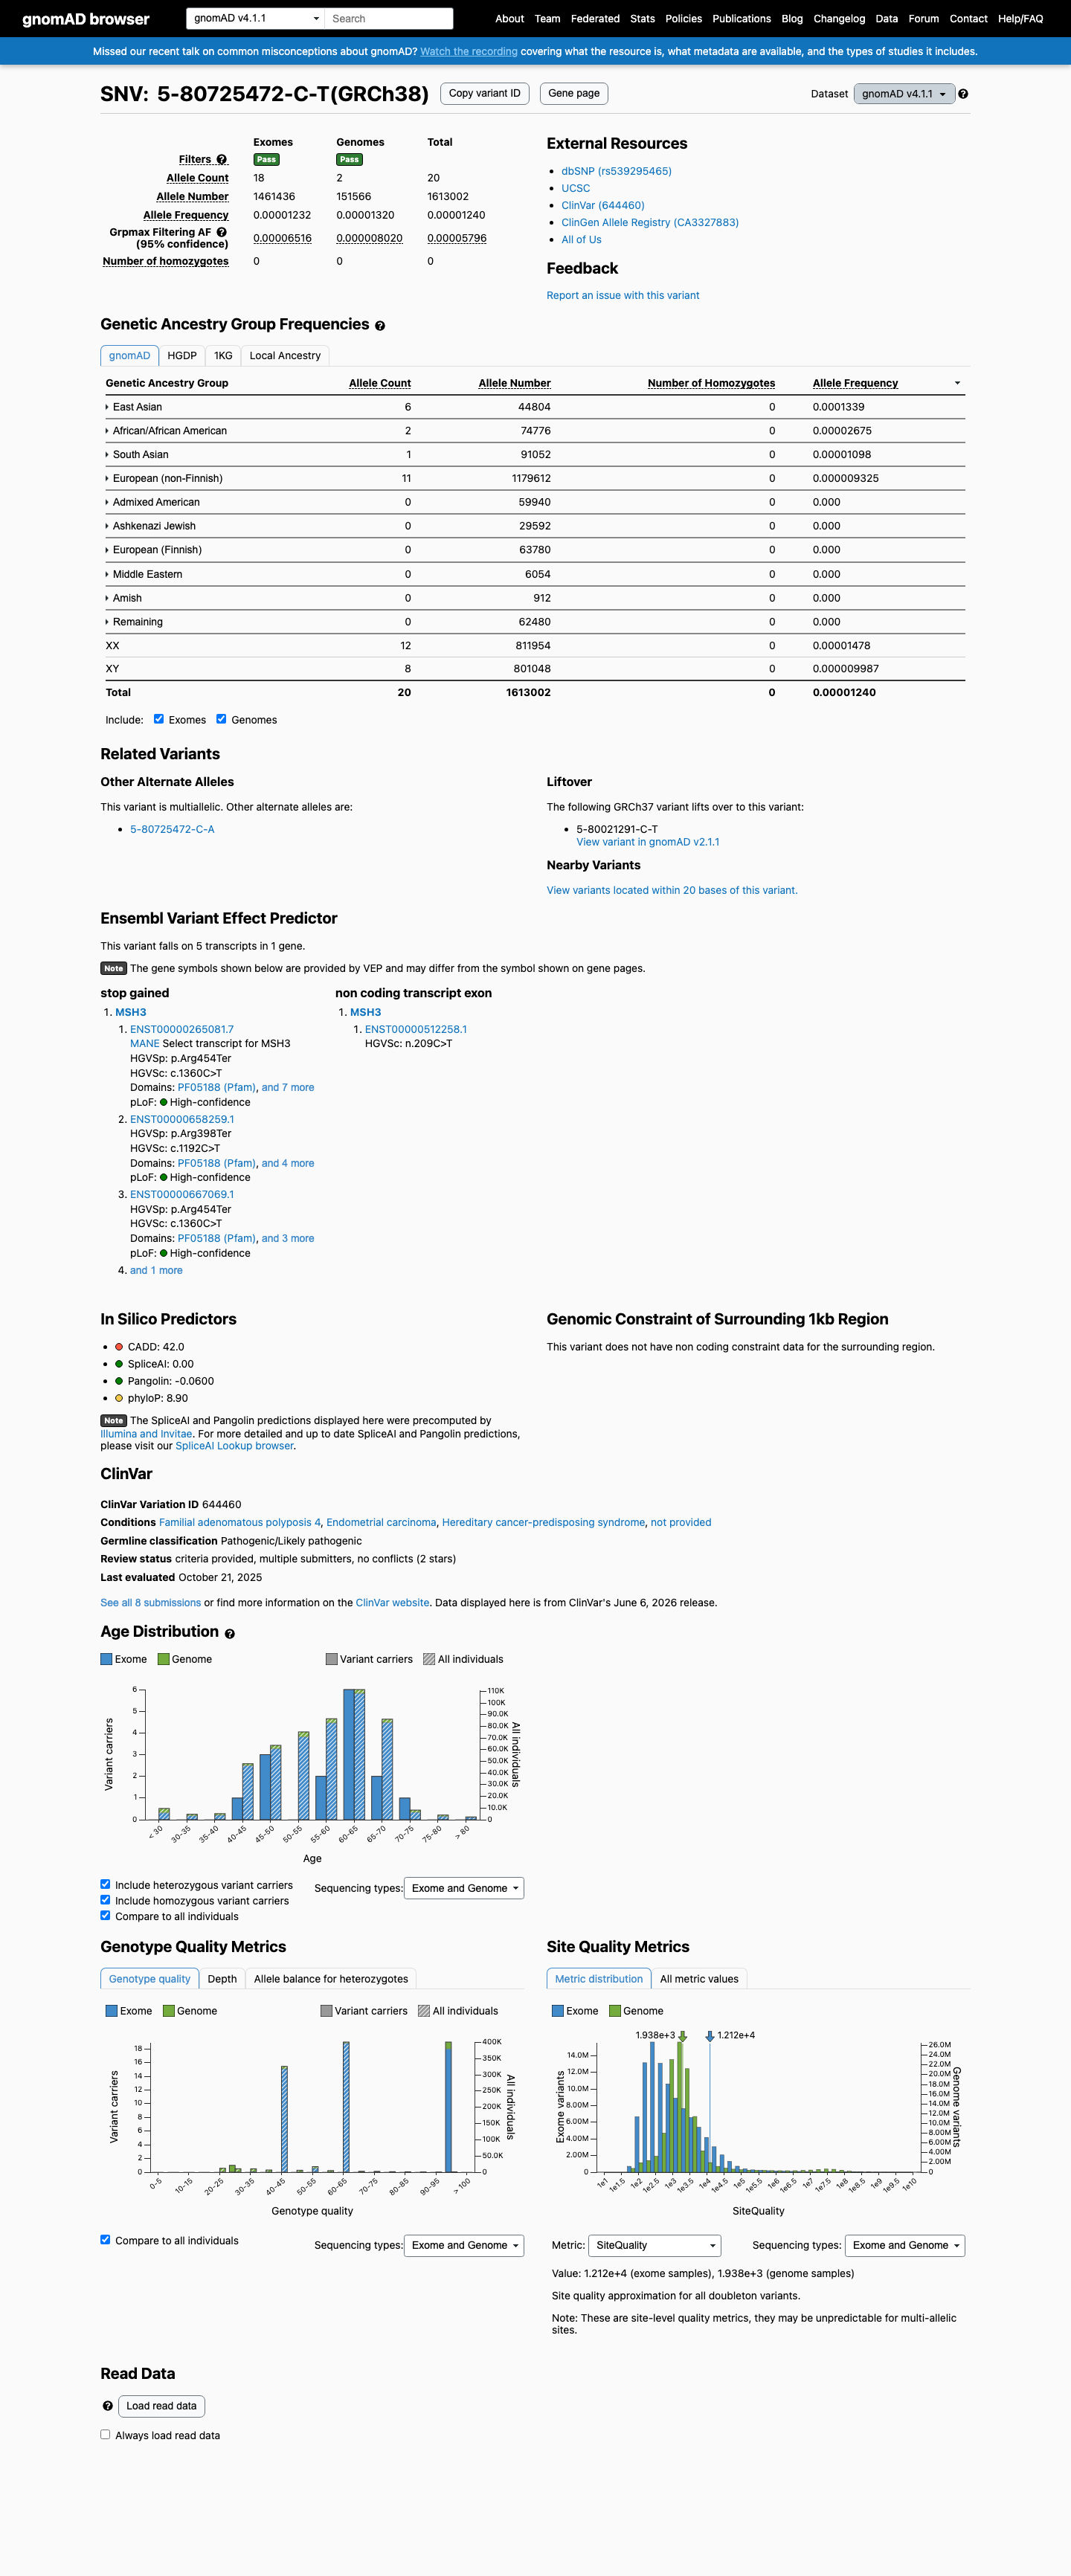
gnomAD v4.1 screenshot
gnomAD v4.1
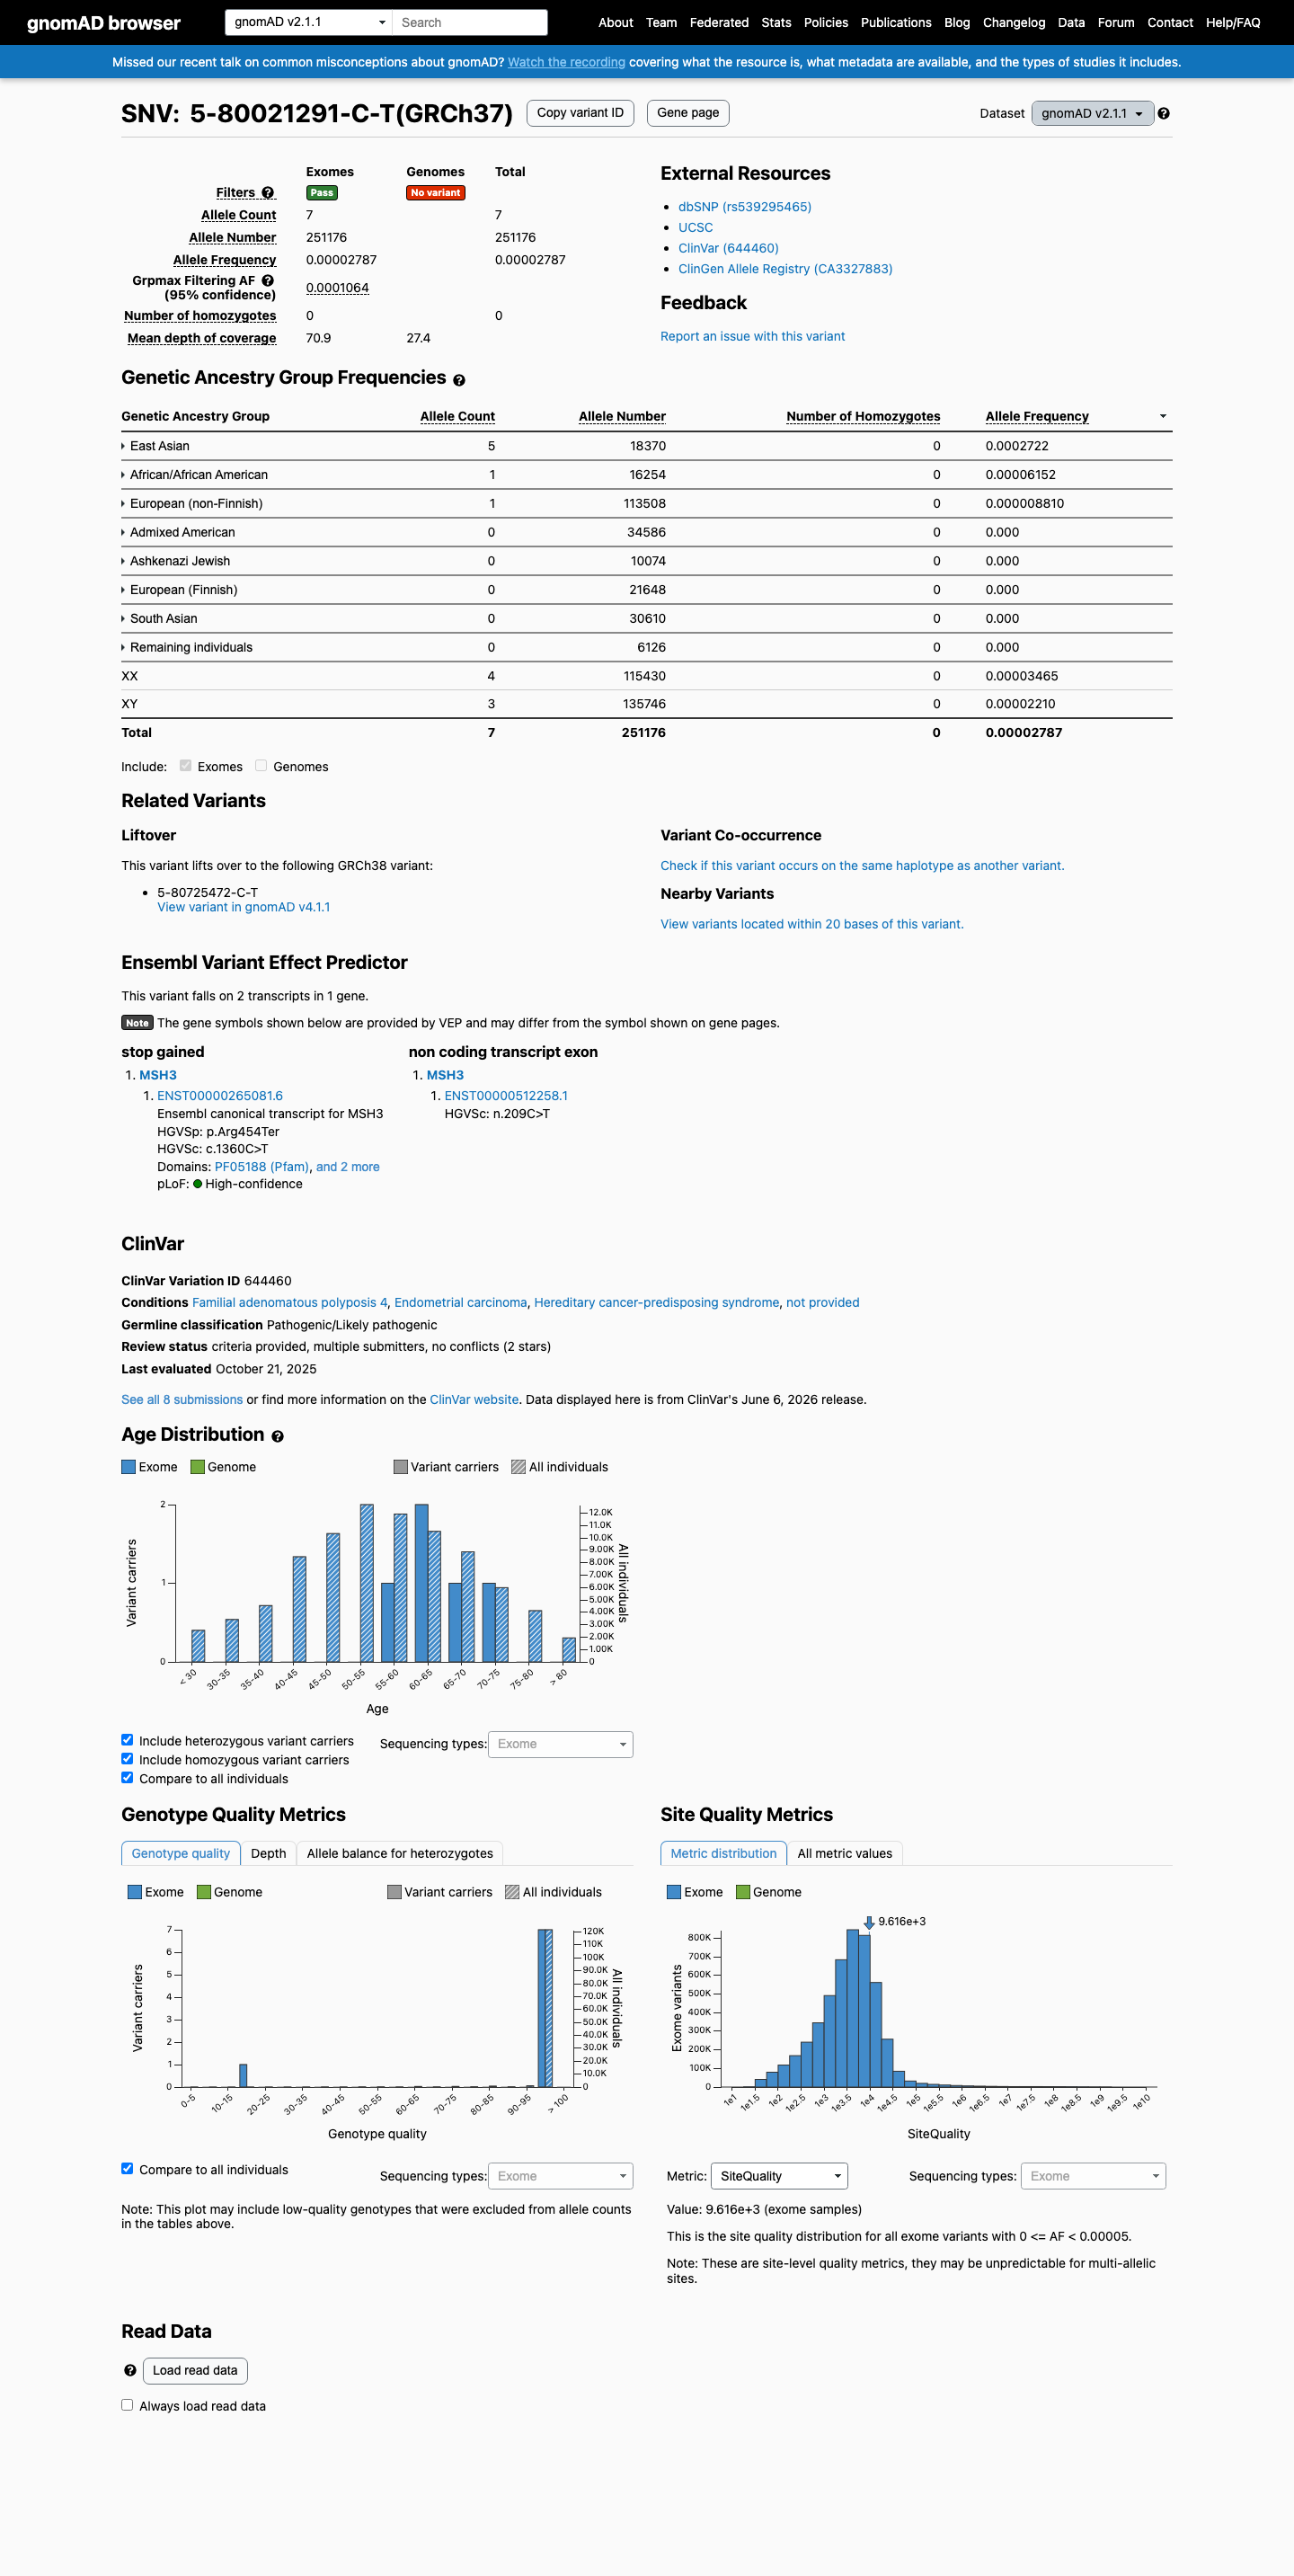
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 1.23992e-05; MAF= 0.00124%, 20/1613002 alleles, homozygotes = 0) and has highest observed frequency in the East Asian population (AF= 0.000133917; MAF= 0.01339%, 6/44804 alleles, homozygotes = 0); grpmax FAF= 5.796e-05.
v2.1
This variant is present in gnomAD v2.1 (AF= 2.78689e-05; MAF= 0.00279%, 7/251176 alleles, homozygotes = 0) and has highest observed frequency in the East Asian population (AF= 0.000272183; MAF= 0.02722%, 5/18370 alleles, homozygotes = 0); grpmax FAF= 0.00010636.
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.0012% · 20 / 1,613,002
0 hom · FAF 0.0058%
East Asian
6 / 44,804
0.013%
African/African American
2 / 74,776
0.0027%
South Asian
1 / 91,052
0.0011%
European (non-Finnish)
11 / 1,179,612
0.00093%
+ 6 not observed (Remaining individuals, Admixed American, European (Finnish), Amish, Middle Eastern, Ashkenazi Jewish)
gnomAD v2.1
0.0028% · 7 / 251,176
0 hom · FAF 0.011%
East Asian
5 / 18,370
0.027%
African/African American
1 / 16,254
0.0062%
European (non-Finnish)
1 / 113,508
0.00088%
+ 5 not observed (Admixed American, Ashkenazi Jewish, European (Finnish), Remaining individuals, South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
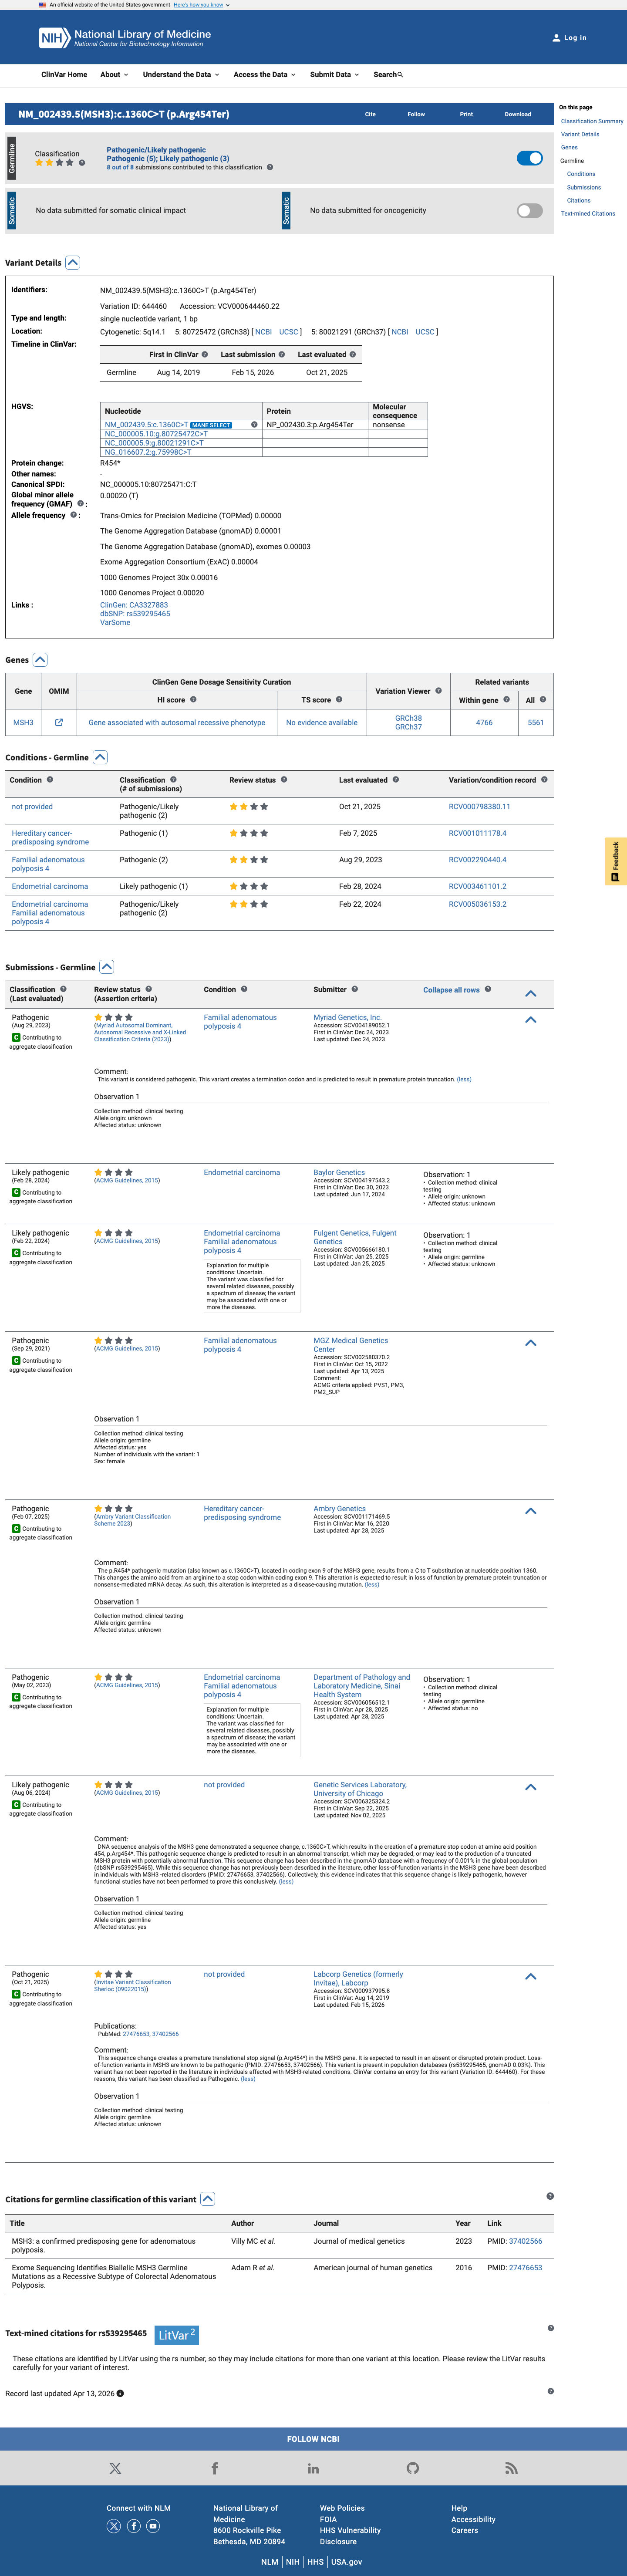
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Pathogenic (5 clinical laboratories) and as Likely pathogenic (3 clinical laboratories). (ClinVarID = 644460)
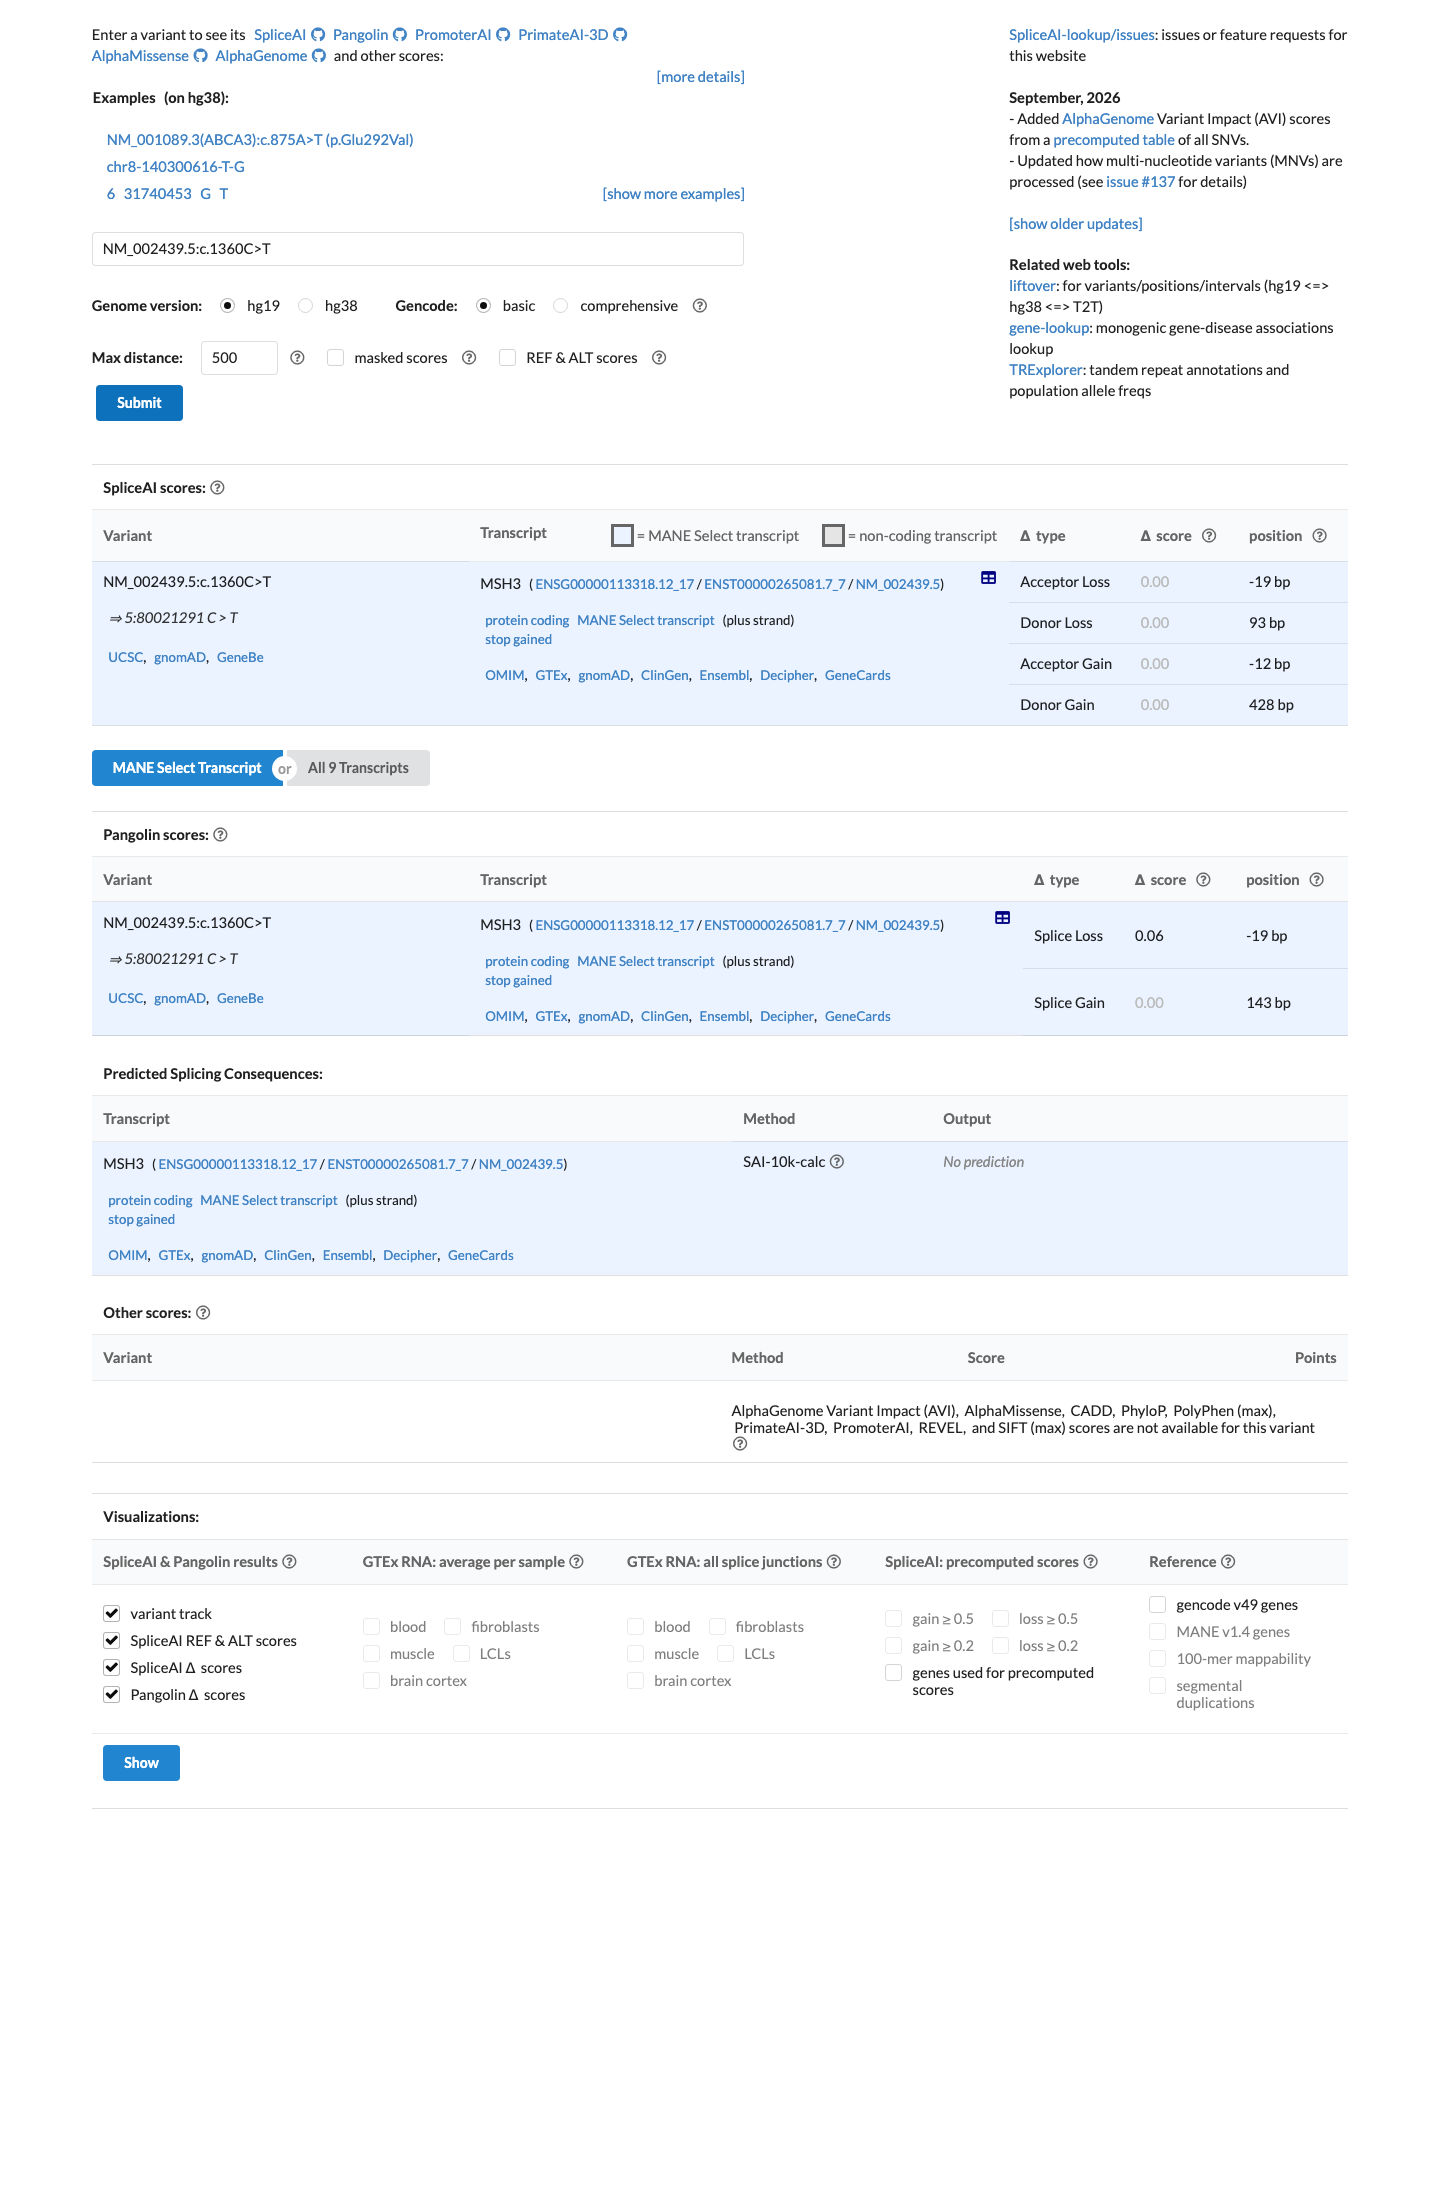
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.00). BayesDel score = 0.61701.
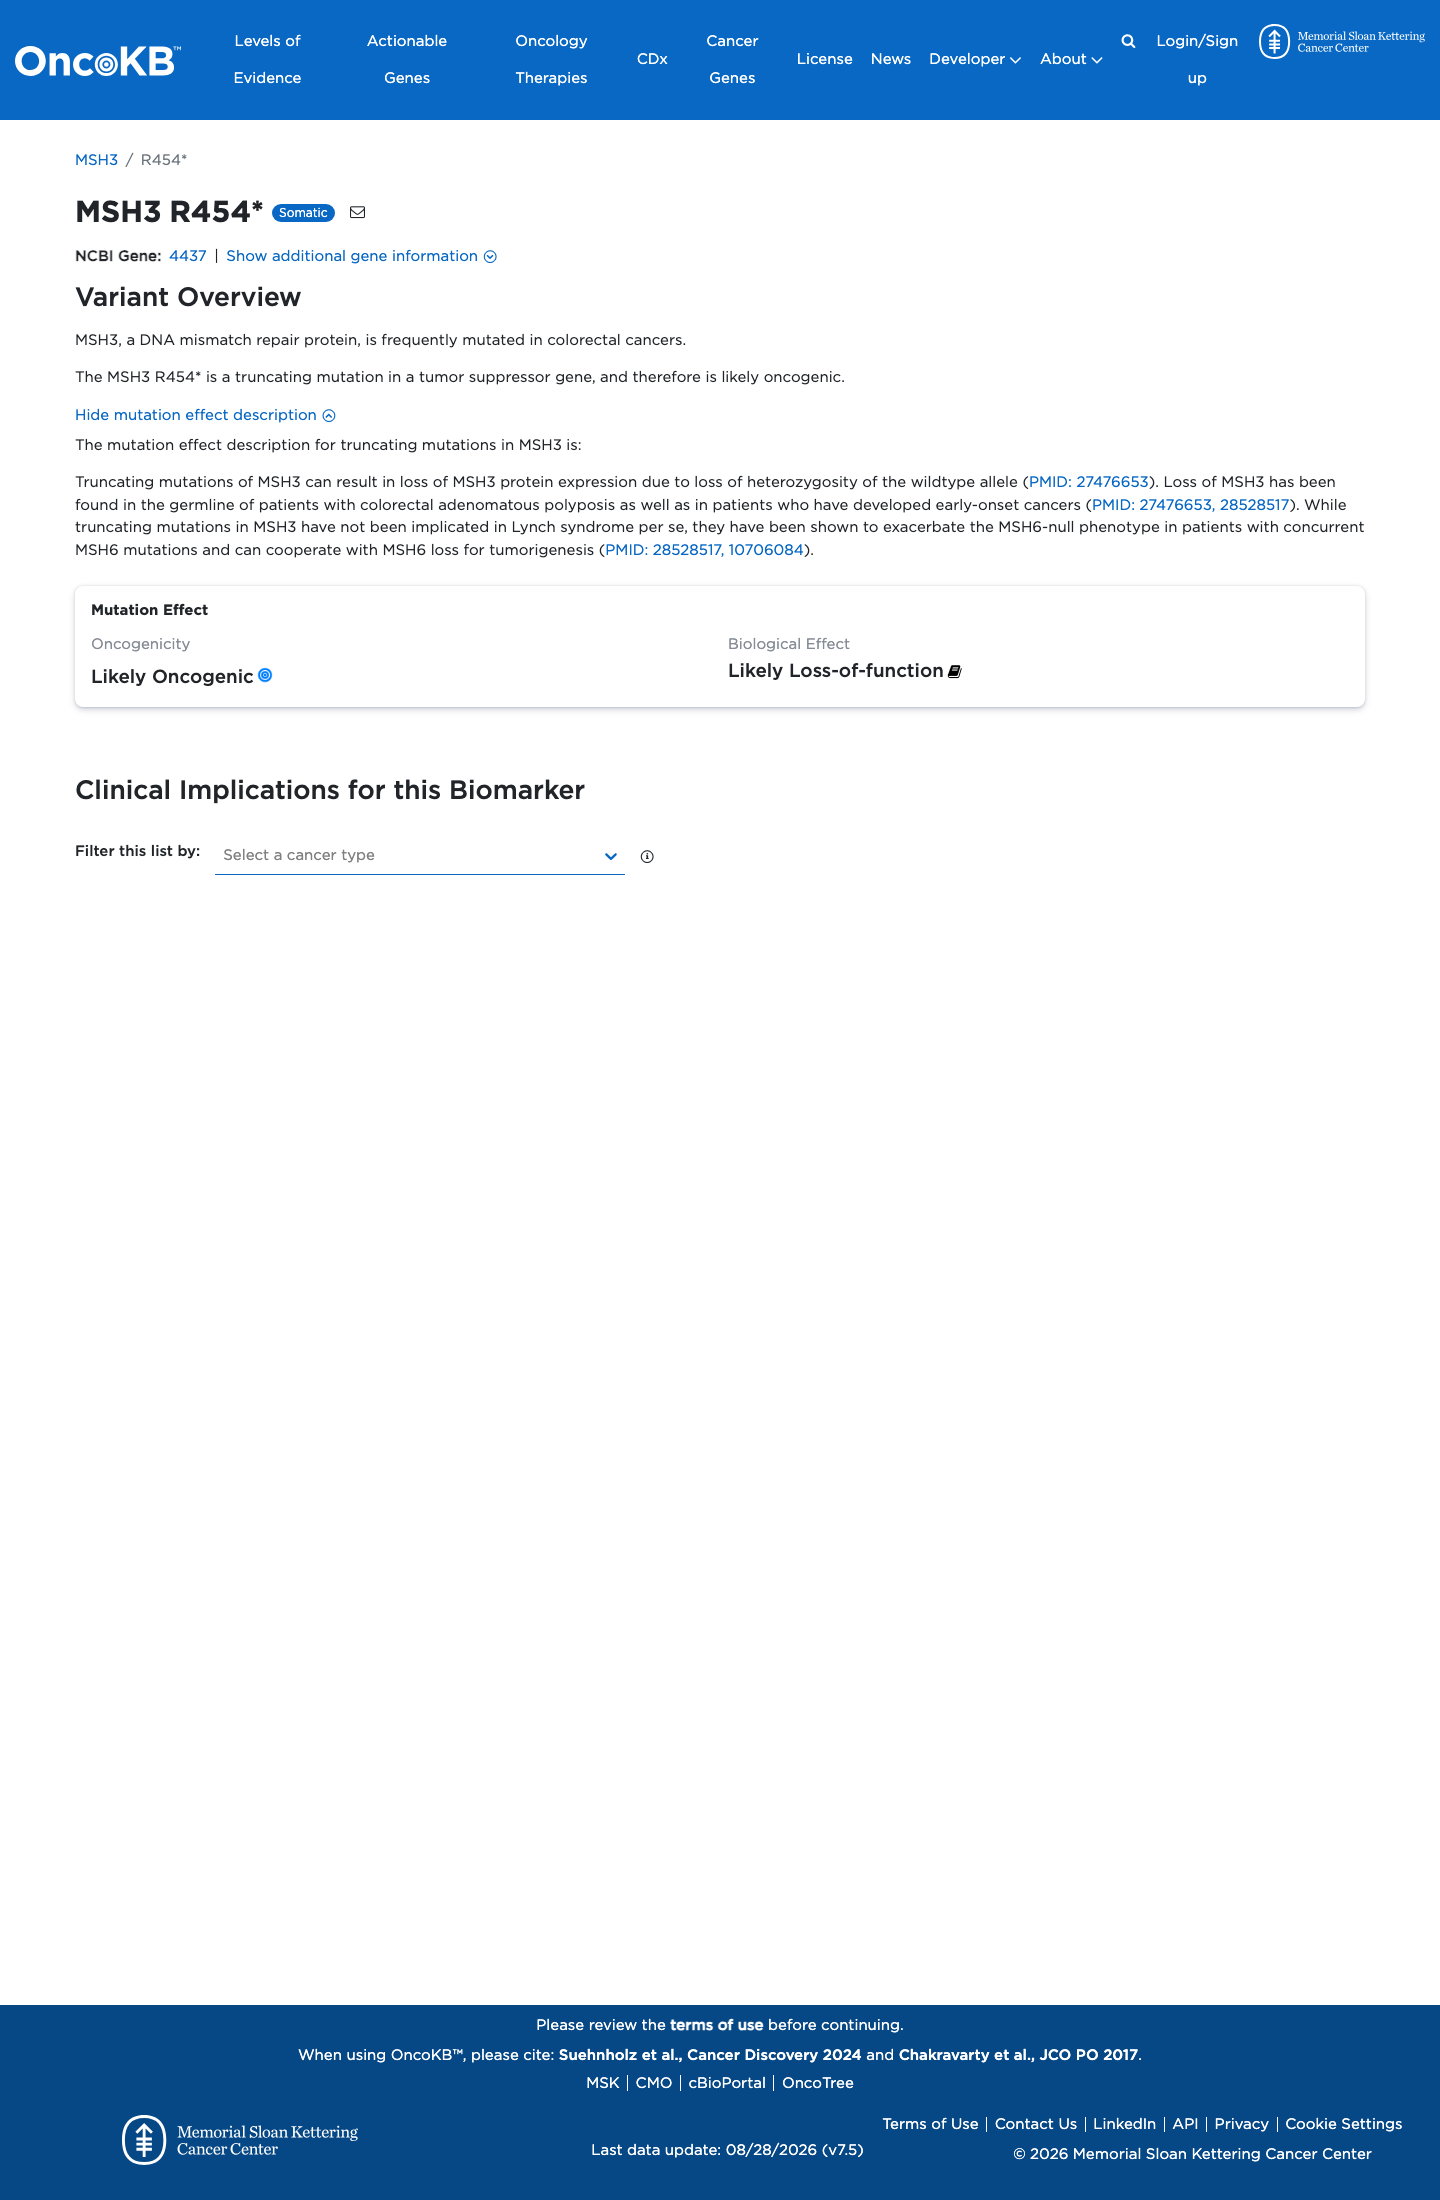
Functional / OncoKB screenshot
Functional Likely Oncogenic
OncoKB identified variant-specific curated literature and context relevant to functional review; biological-effect context: Likely Loss-of-function; curated oncogenicity label: Likely Oncogenic.
OncoKB ↗
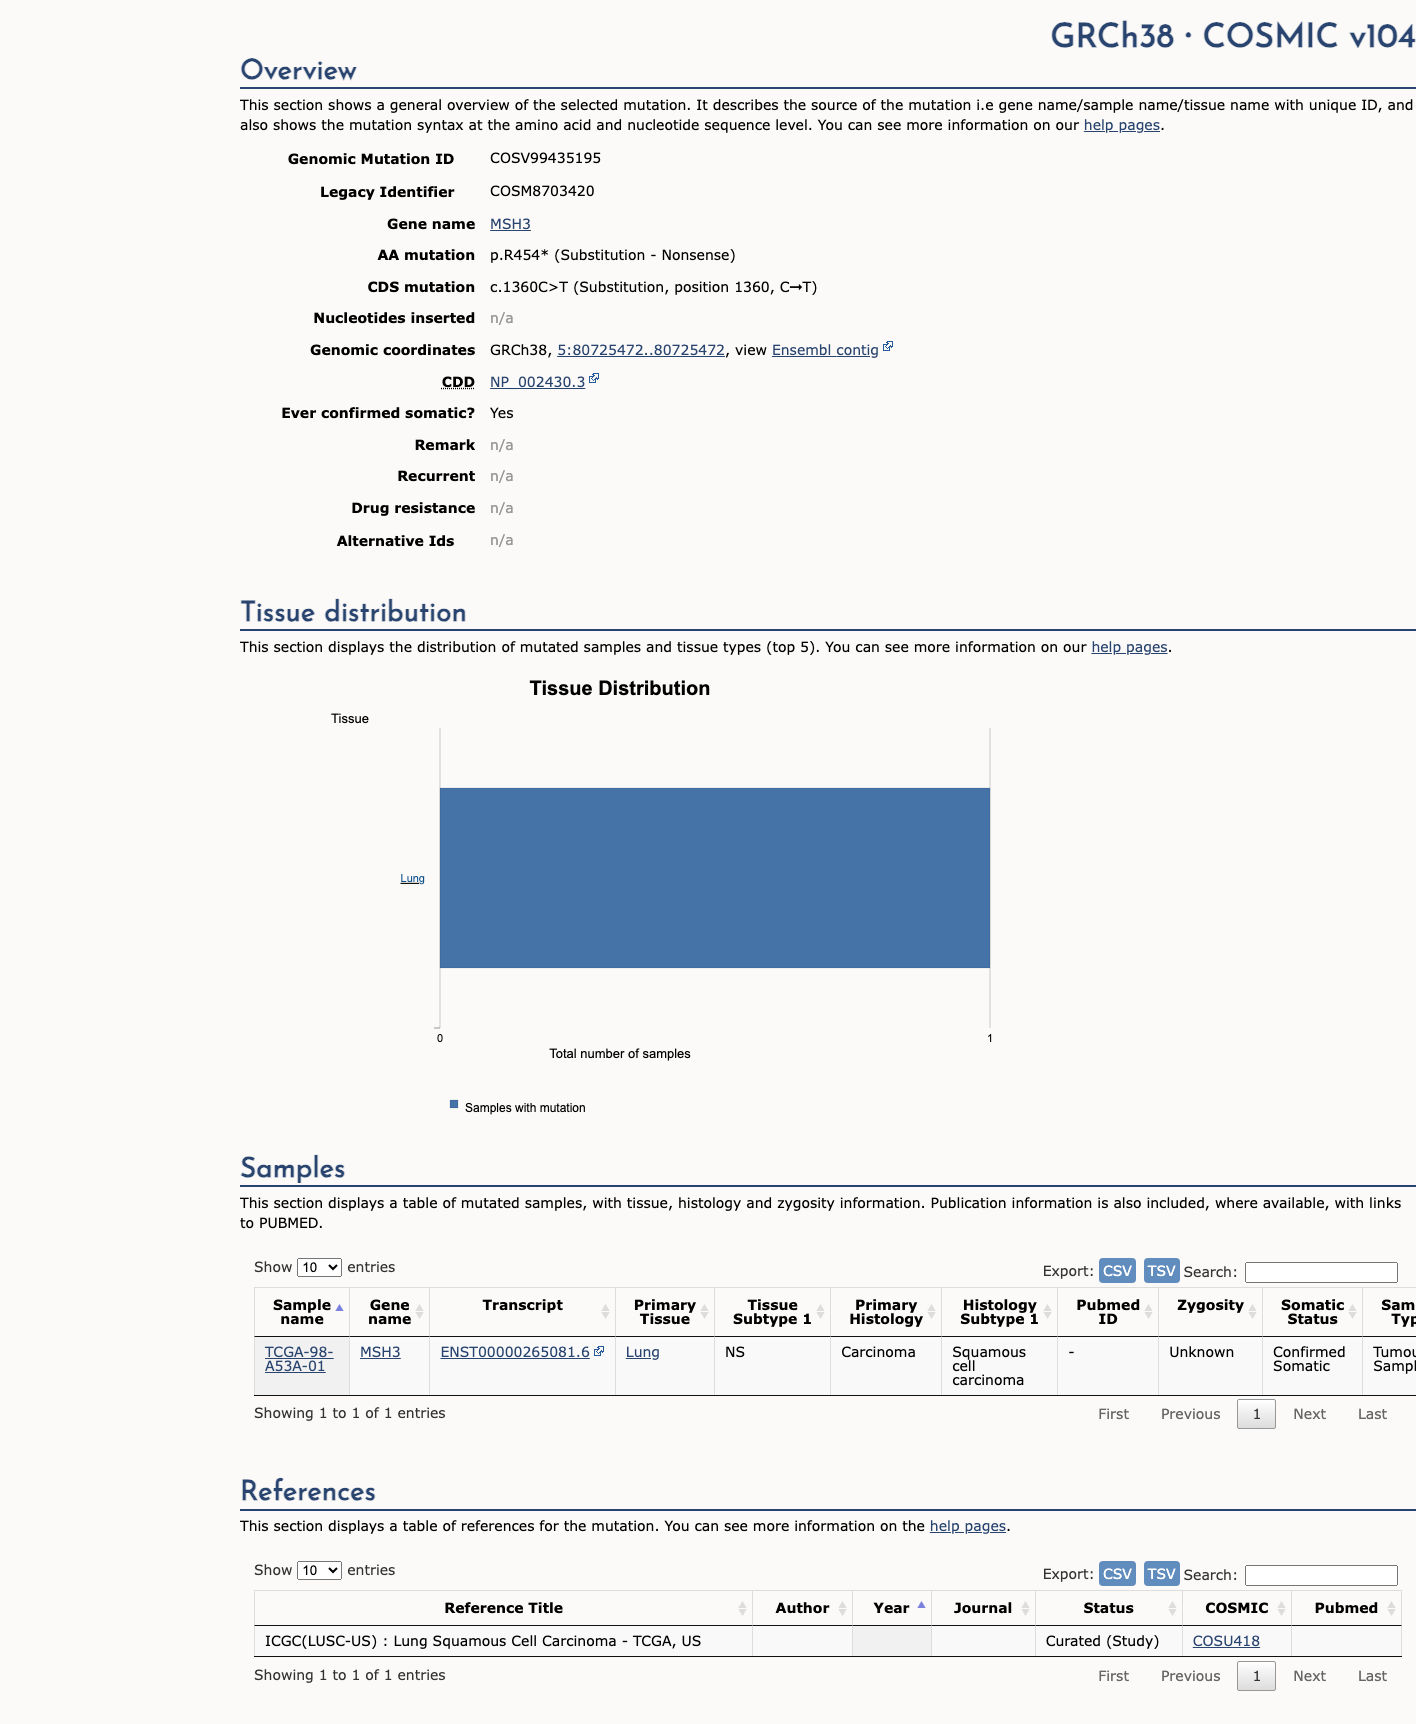
COSMIC screenshot
COSMIC
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has previously been reported in somatic cancers (COSMIC; COSV99435195, n = 1 times).
Hotspots
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
4papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 4 further PMIDs triaged but not cited — see Sources & references.
Rule & framework references · cited for criterion definitions, not variant evidence
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.
27476653 ↗ Exome Sequencing Identifies Biallelic MSH3 Germline Mutations as a Recessive Subtype of Colorectal Adenomatous Polyposis.
28528517 ↗ Loss of MSH2 and MSH6 due to heterozygous germline defects in MSH3 and MSH6.
37402566 ↗ MSH3: a confirmed predisposing gene for adenomatous polyposis.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 4 PMIDs not cited in assessment
10706084 ↗ The DNA mismatch repair genes Msh3 and Msh6 cooperate in intestinal tumor suppression. ONCOKB
24493721 ↗ American Society of Clinical Oncology Expert Statement: collection and use of a cancer family history for oncology providers. CLINVAR
25394175 ↗ A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment. CLINVAR
24905773 ↗ Endometrial cancer: a review and current management strategies: part I. CLINVAR