Back
NM_002691.4:c.353C>T
p.Ser118Phe · POLD1
ACMG/AMP
0%
complete
Final classification
VUS
BP4
POLD1
c.353C>T
p.Ser118Phe
missense · exon 4

POLD1 encodes the catalytic subunit of DNA polymerase delta, an enzyme that carries both DNA synthesis and proofreading (3' to 5' exonuclease) activities and is essential for accurate DNA replication and repair. Germline mutations in its exonuclease domain cause polyposis and predispose people to colorectal, endometrial, and possibly brain cancers. In cancer, POLD1 defects impair replication fidelity, leading to the accumulation of many mutations (an ultra-mutated phenotype) that may make tumors more responsive to immunotherapy, though somatic POLD1 mutations are rare.

This variant

POLD1 encodes the catalytic DNA polymerase delta subunit, whose proofreading activity maintains replication fidelity and whose germline exonuclease-domain defects predispose to polyposis and several cancers.

Transcript
NM_002691.4
HGVS · transcript:coding
NM_002691.4:c.353C>T
GRCh38
chr19:50401814 C>T
GRCh37
chr19:50905071 C>T
VUS: BP4 (supporting) alone does not satisfy the generic ACMG/AMP threshold for Likely Benign or Benign.
Classification rationale
BP4 VUS
POLD1 c.353C>T missense · exon 4

BP4 supporting: REVEL 0.099 is below the benign computational threshold. VUS: BP4 alone is insufficient for the generic ACMG/AMP Likely Benign or Benign thresholds.

BP4 → VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_002691.4 · variants mapped to exon structure
POLD1 NM_002691.4
Fetching transcript structure from UCSC…
Applied criteria · 1 applied · 23 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 1
Strength Supporting Moderate Strong Very strong
BP4 supporting Benign
Met at supporting: REVEL 0.099 is below the BP4 supporting threshold of <=0.29 for missense variants.
The variant is missense, NP_002682.2:p.(Ser118Phe), so REVEL is the applicable computational path and SpliceAI is excluded.REVEL score is 0.099, meeting BP4 supporting <=0.29 but not the moderate <=0.183 or strong <=0.016 thresholds under the ClinGen SVI REVEL calibration (Pejaver et al. 2022, PMID:36413997).BayesDel has no generic ACMG-strength calibration and was not used for BP4.
Assessed · not applied · 8 not met · 15 not assessed
Pathogenic
PS1 Not assessed: no validated pathogenic POLD1 substitution producing the same p.Ser118Phe amino-acid change was identified.
PS2 Not assessed: no documented affected proband with confirmed absence of the variant in both tested biological parents.
PS3 Not assessed: no variant-specific validated functional assay, assay controls, or quantitative biological readout was reported for POLD1 p.Ser118Phe.
PS4 Not assessed: no variant-specific case-control enrichment, odds ratio, likelihood ratio, or affected-versus-control prevalence data are available.
PM1 Not met: POLD1 S118 is not in a documented hotspot, and no approved critical-domain evidence places residue 118 in a PM1 region.
PM2 Not met: aggregate AF is below 0.0001, but ancestry-specific frequencies reach 0.000696185 in gnomAD v2.1 and exceed the PM2 threshold.
PM3 Not assessed: no affected-proband biallelic observation or verified pathogenic variant in trans is documented for this POLD1 variant.
PM5 Not assessed: no validated pathogenic missense comparator at POLD1 residue Ser118 was available for PM5.
PM6 Not assessed: no suspected de novo observation with compatible phenotype and parental testing or parentage information is reported.
PP1 Not assessed: no informative affected-relative or unaffected-relative genotypes and no segregation meioses are documented.
PP2 Not assessed: no gene-specific POLD1 evidence establishes both a dominant missense mechanism and a low benign-missense rate for PP2.
PP3 Not met: REVEL 0.099 is below the PP3 supporting threshold of >=0.644 for missense variants.
PP4 Not assessed: no carrier phenotype or clinical diagnosis is provided to evaluate specificity for a POLD1-related disorder.
PP5 Not met: ClinVar has zero expert-panel submissions for the exact variant and no qualifying Pathogenic or Likely pathogenic expert-panel classification.
Benign
BA1 Not met: the highest default all-comers subpopulation frequency is 0.000696185, far below the 0.05 BA1 threshold.
BS1 Not met: the highest default all-comers subpopulation frequency, 0.000696185, is about 14-fold below the 0.01 BS1 threshold.
BS2 Not met: all available datasets show zero homozygotes, and gnomAD provides no phenotype or age data proving any carrier is a healthy adult.
BS3 Not assessed: no variant-specific validated assay demonstrated preserved POLD1 function with appropriate controls or a quantitative benign-effect result.
BS4 Not assessed: no tested unaffected relatives with documented non-segregation and reliable phenotype information are reported.
BP1 Not assessed: POLD1 is missense here, but predominant truncating-variant disease mechanism has not been established for BP1.
BP2 Not assessed: no documented co-occurrence or phase result shows this variant with another pathogenic or likely pathogenic variant.
BP5 Not assessed: no alternative molecular diagnosis or evidence that another variant explains the patient's phenotype is documented.
BP6 Not met: ClinVar has zero expert-panel submissions, so its ordinary Likely benign laboratory assertions cannot trigger BP6.
N/A · 4 PVS1 · PM4 · BP3 · BP7
Research & evidence
Population frequency
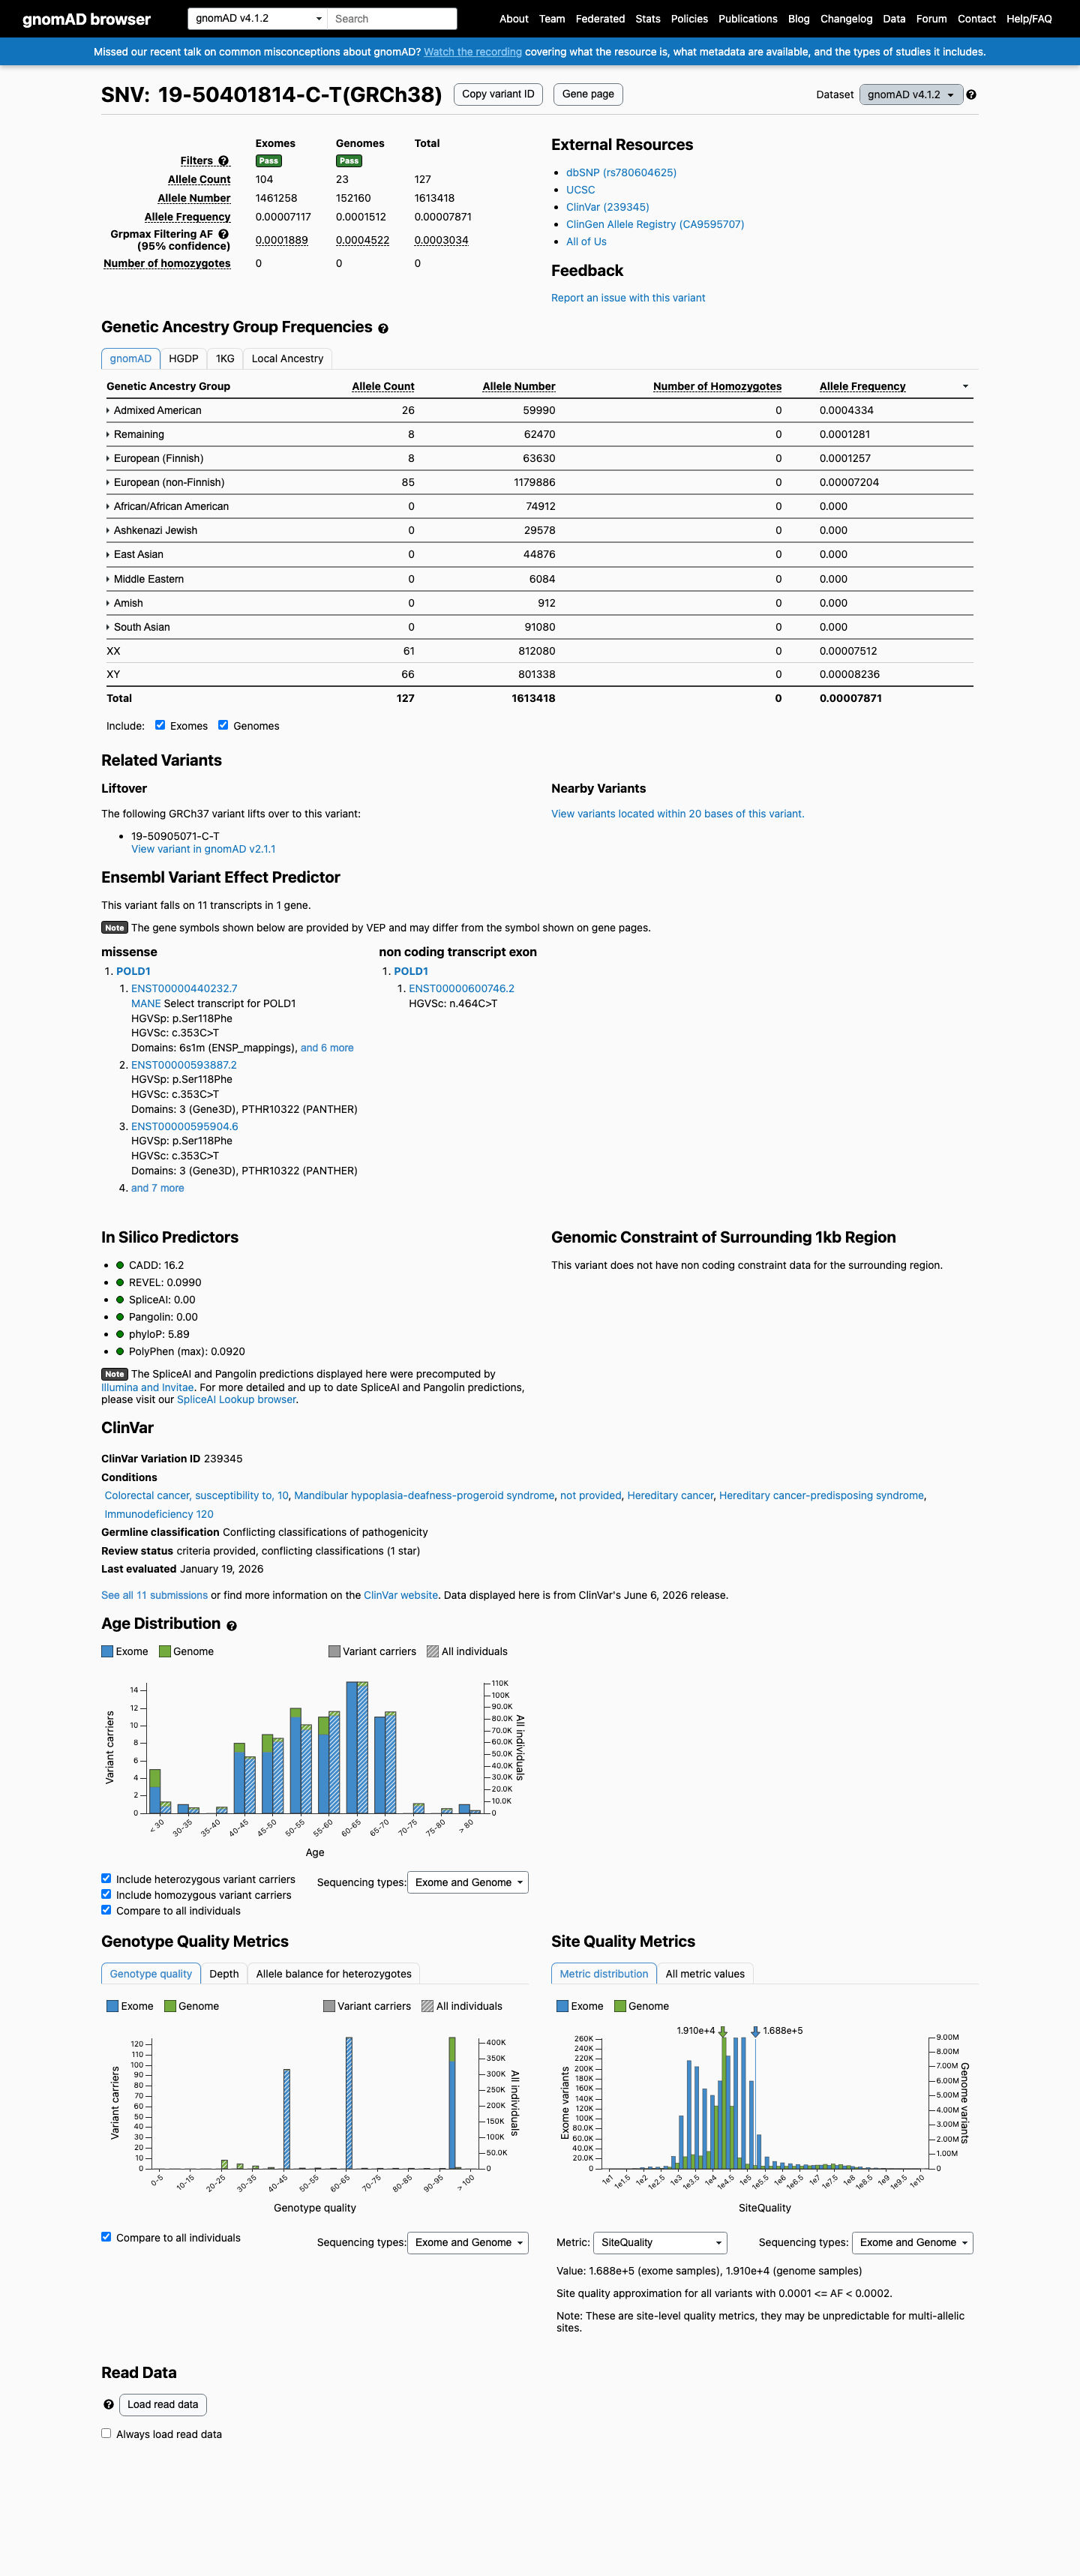
gnomAD v4.1 screenshot
gnomAD v4.1
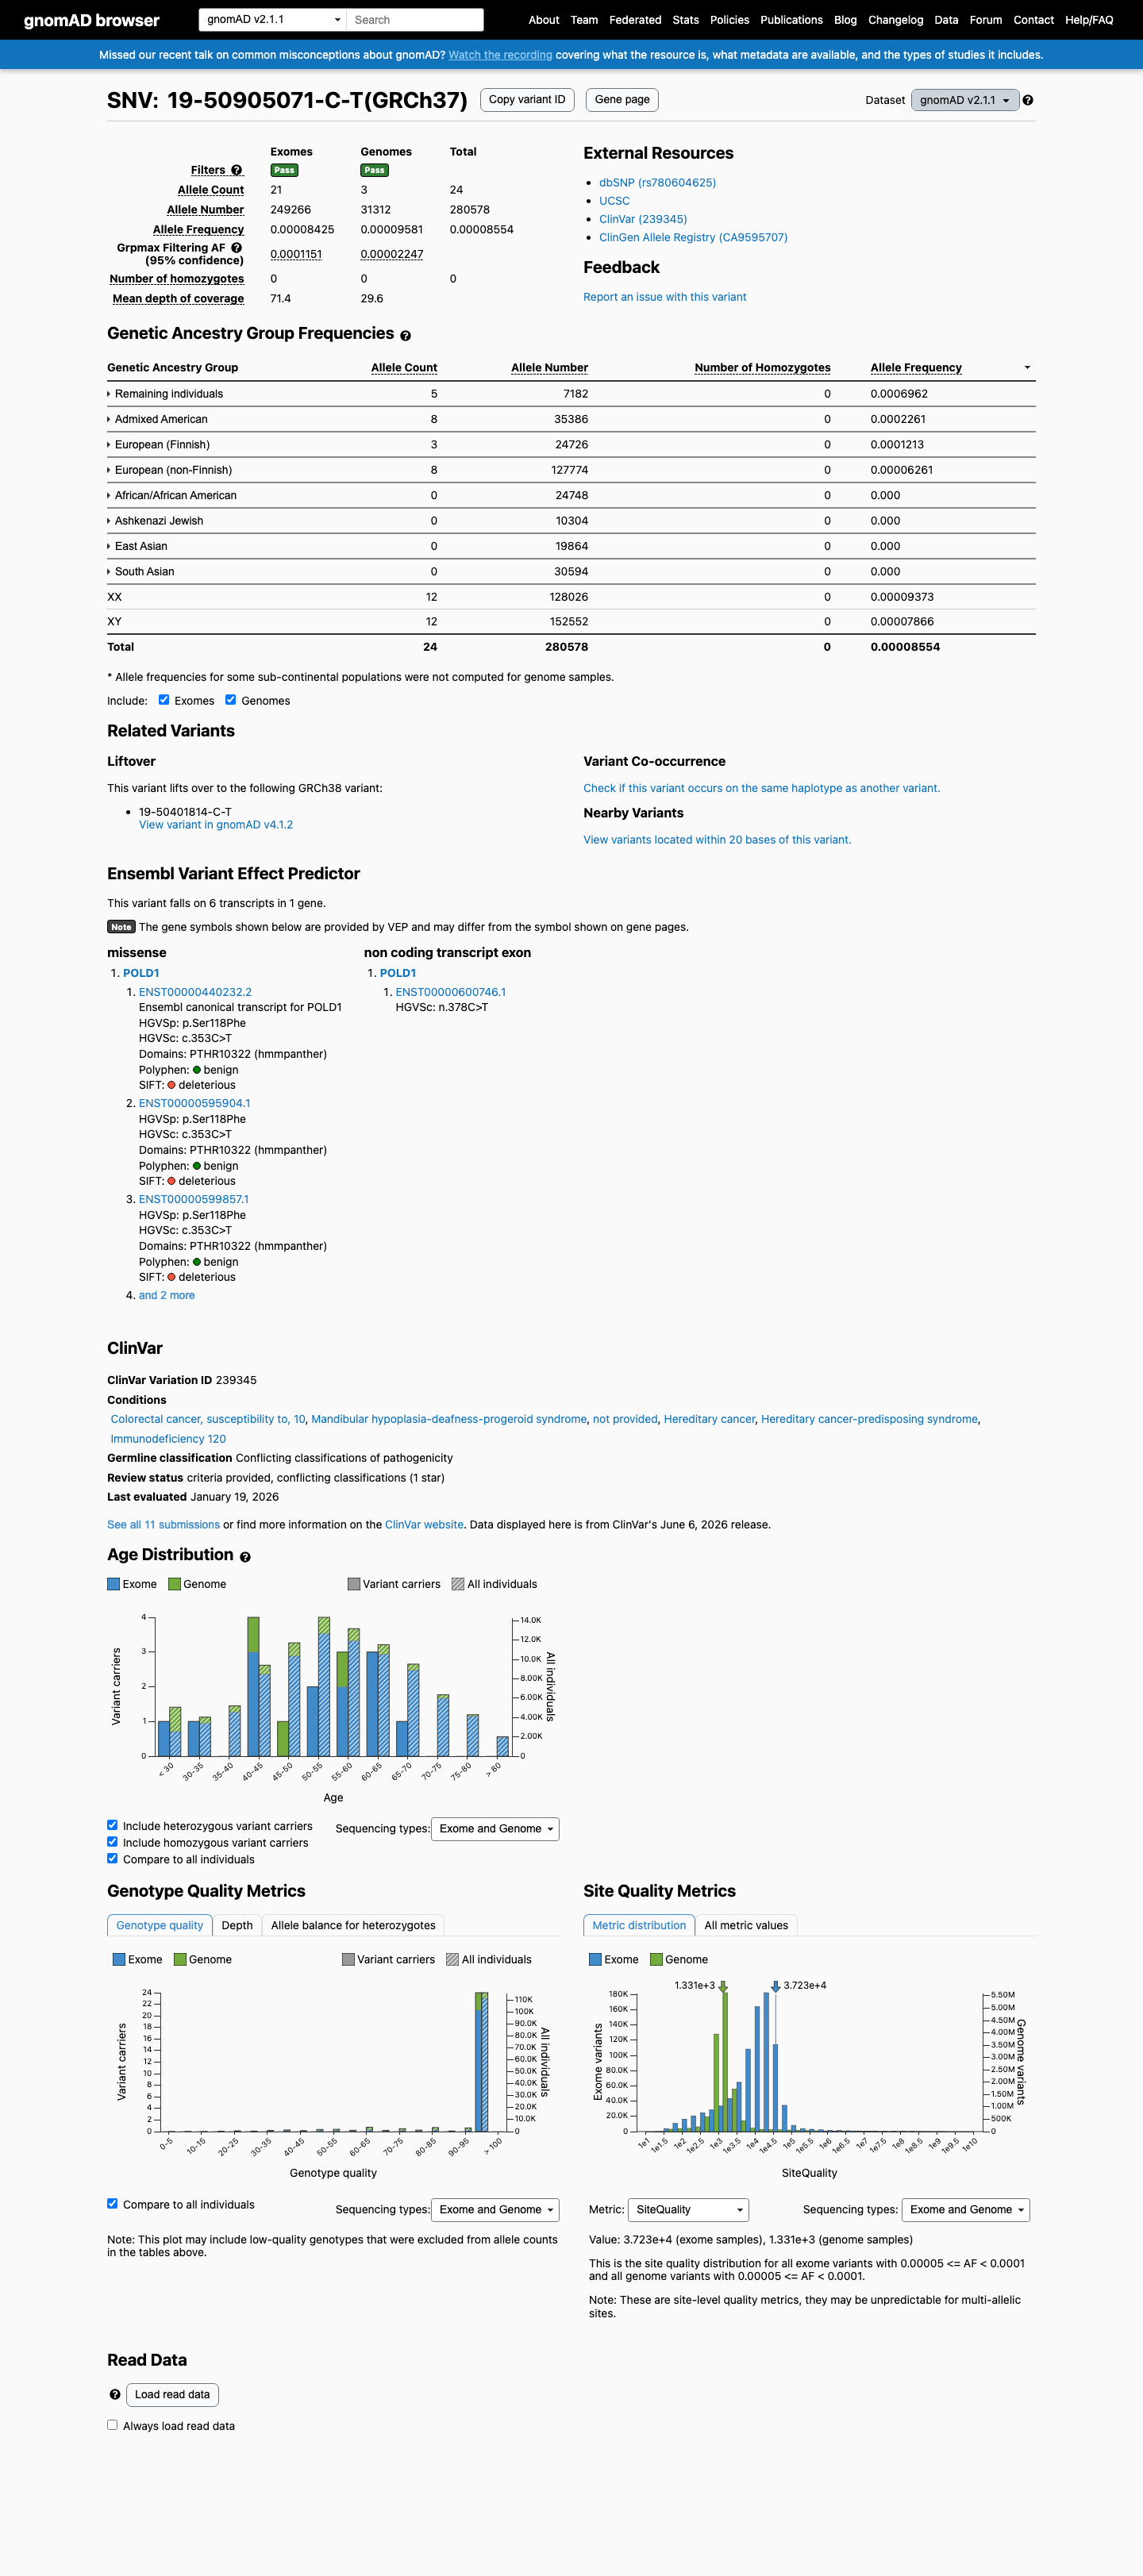
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 7.87149e-05; MAF= 0.00787%, 127/1613418 alleles, homozygotes = 0) and has highest observed frequency in the Admixed American population (AF= 0.000433406; MAF= 0.04334%, 26/59990 alleles, homozygotes = 0); grpmax FAF= 0.0003034.
v2.1
This variant is present in gnomAD v2.1 (AF= 8.55377e-05; MAF= 0.00855%, 24/280578 alleles, homozygotes = 0) and has highest observed frequency in the Remaining individuals population (AF= 0.000696185; MAF= 0.06962%, 5/7182 alleles, homozygotes = 0); grpmax FAF= 0.00011513.
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.0079% · 127 / 1,613,418
0 hom · FAF 0.03%
Admixed American
26 / 59,990
0.043%
Remaining individuals
8 / 62,470
0.013%
European (Finnish)
8 / 63,630
0.013%
European (non-Finnish)
85 / 1,179,886
0.0072%
+ 6 not observed (Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American)
gnomAD v2.1
0.0086% · 24 / 280,578
0 hom · FAF 0.012%
Remaining individuals
5 / 7,182
0.07%
Admixed American
8 / 35,386
0.023%
European (Finnish)
3 / 24,726
0.012%
European (non-Finnish)
8 / 127,774
0.0063%
+ 4 not observed (African/African American, Ashkenazi Jewish, East Asian, South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
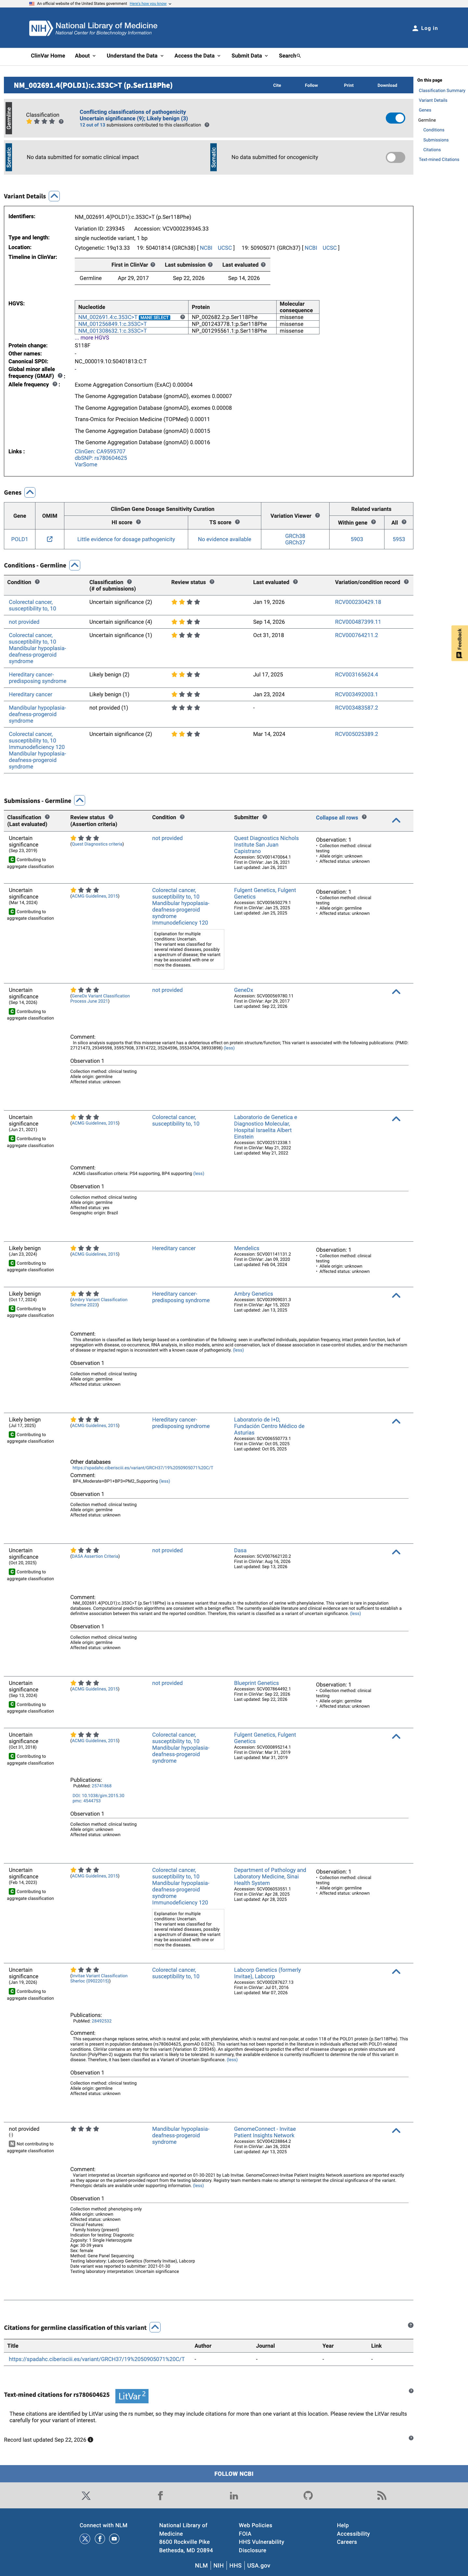
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Uncertain significance (9 clinical laboratories) and as Likely benign (3 clinical laboratories). (ClinVarID = 239345)
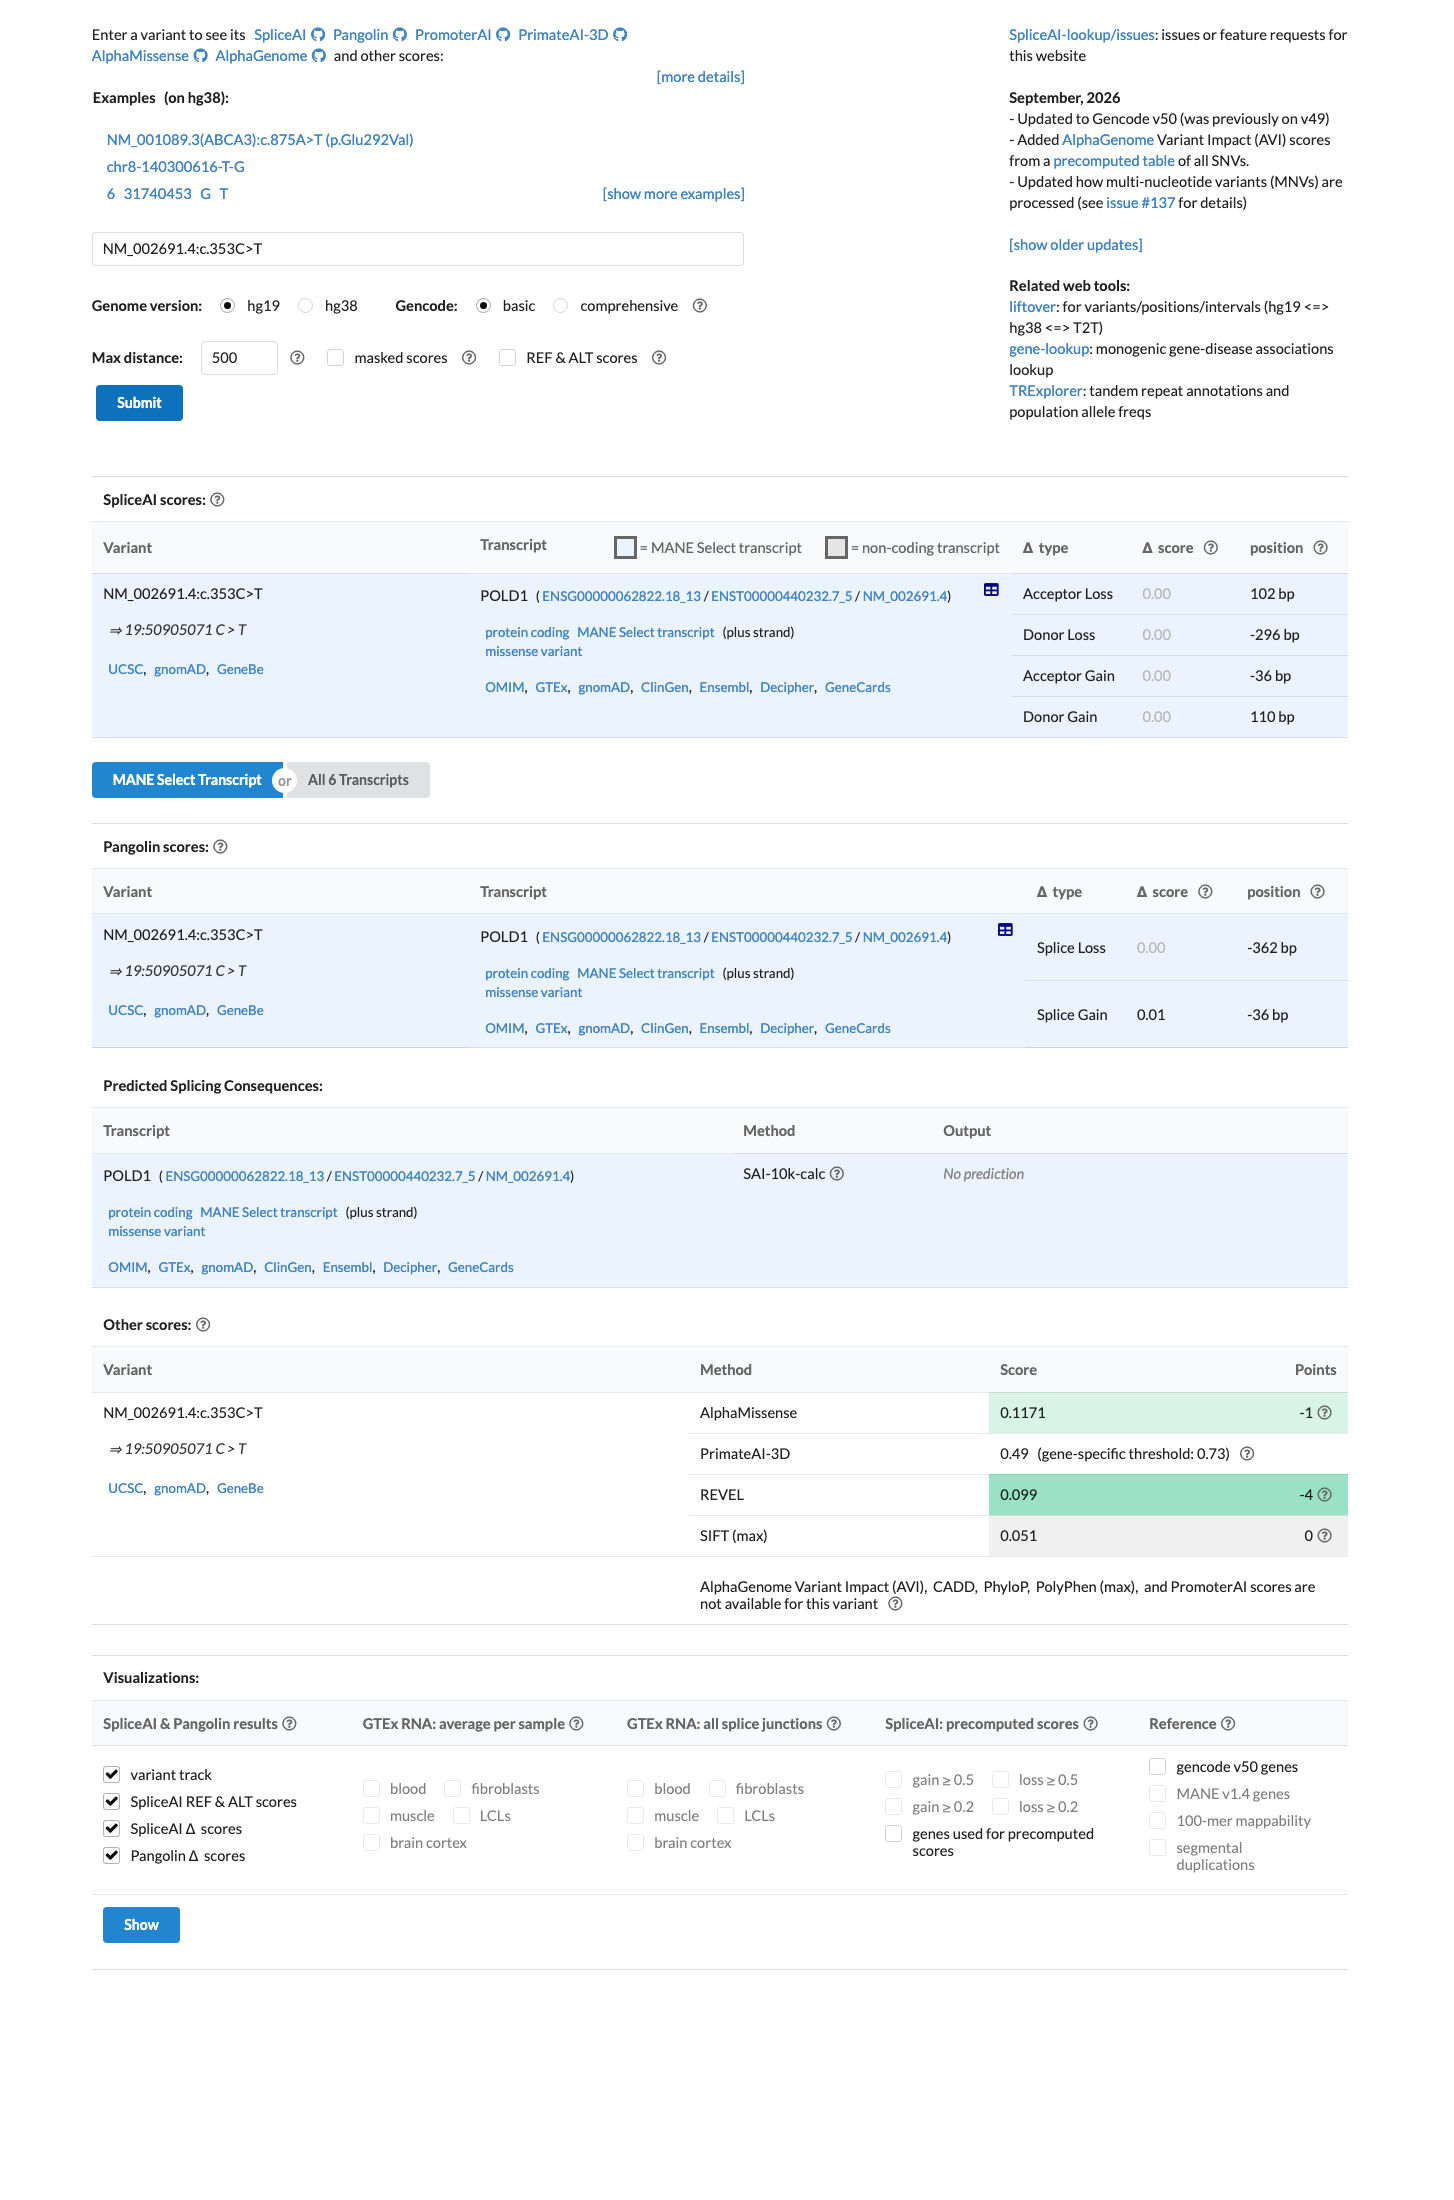
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.00). REVEL score = 0.099. BayesDel score = -0.42693.
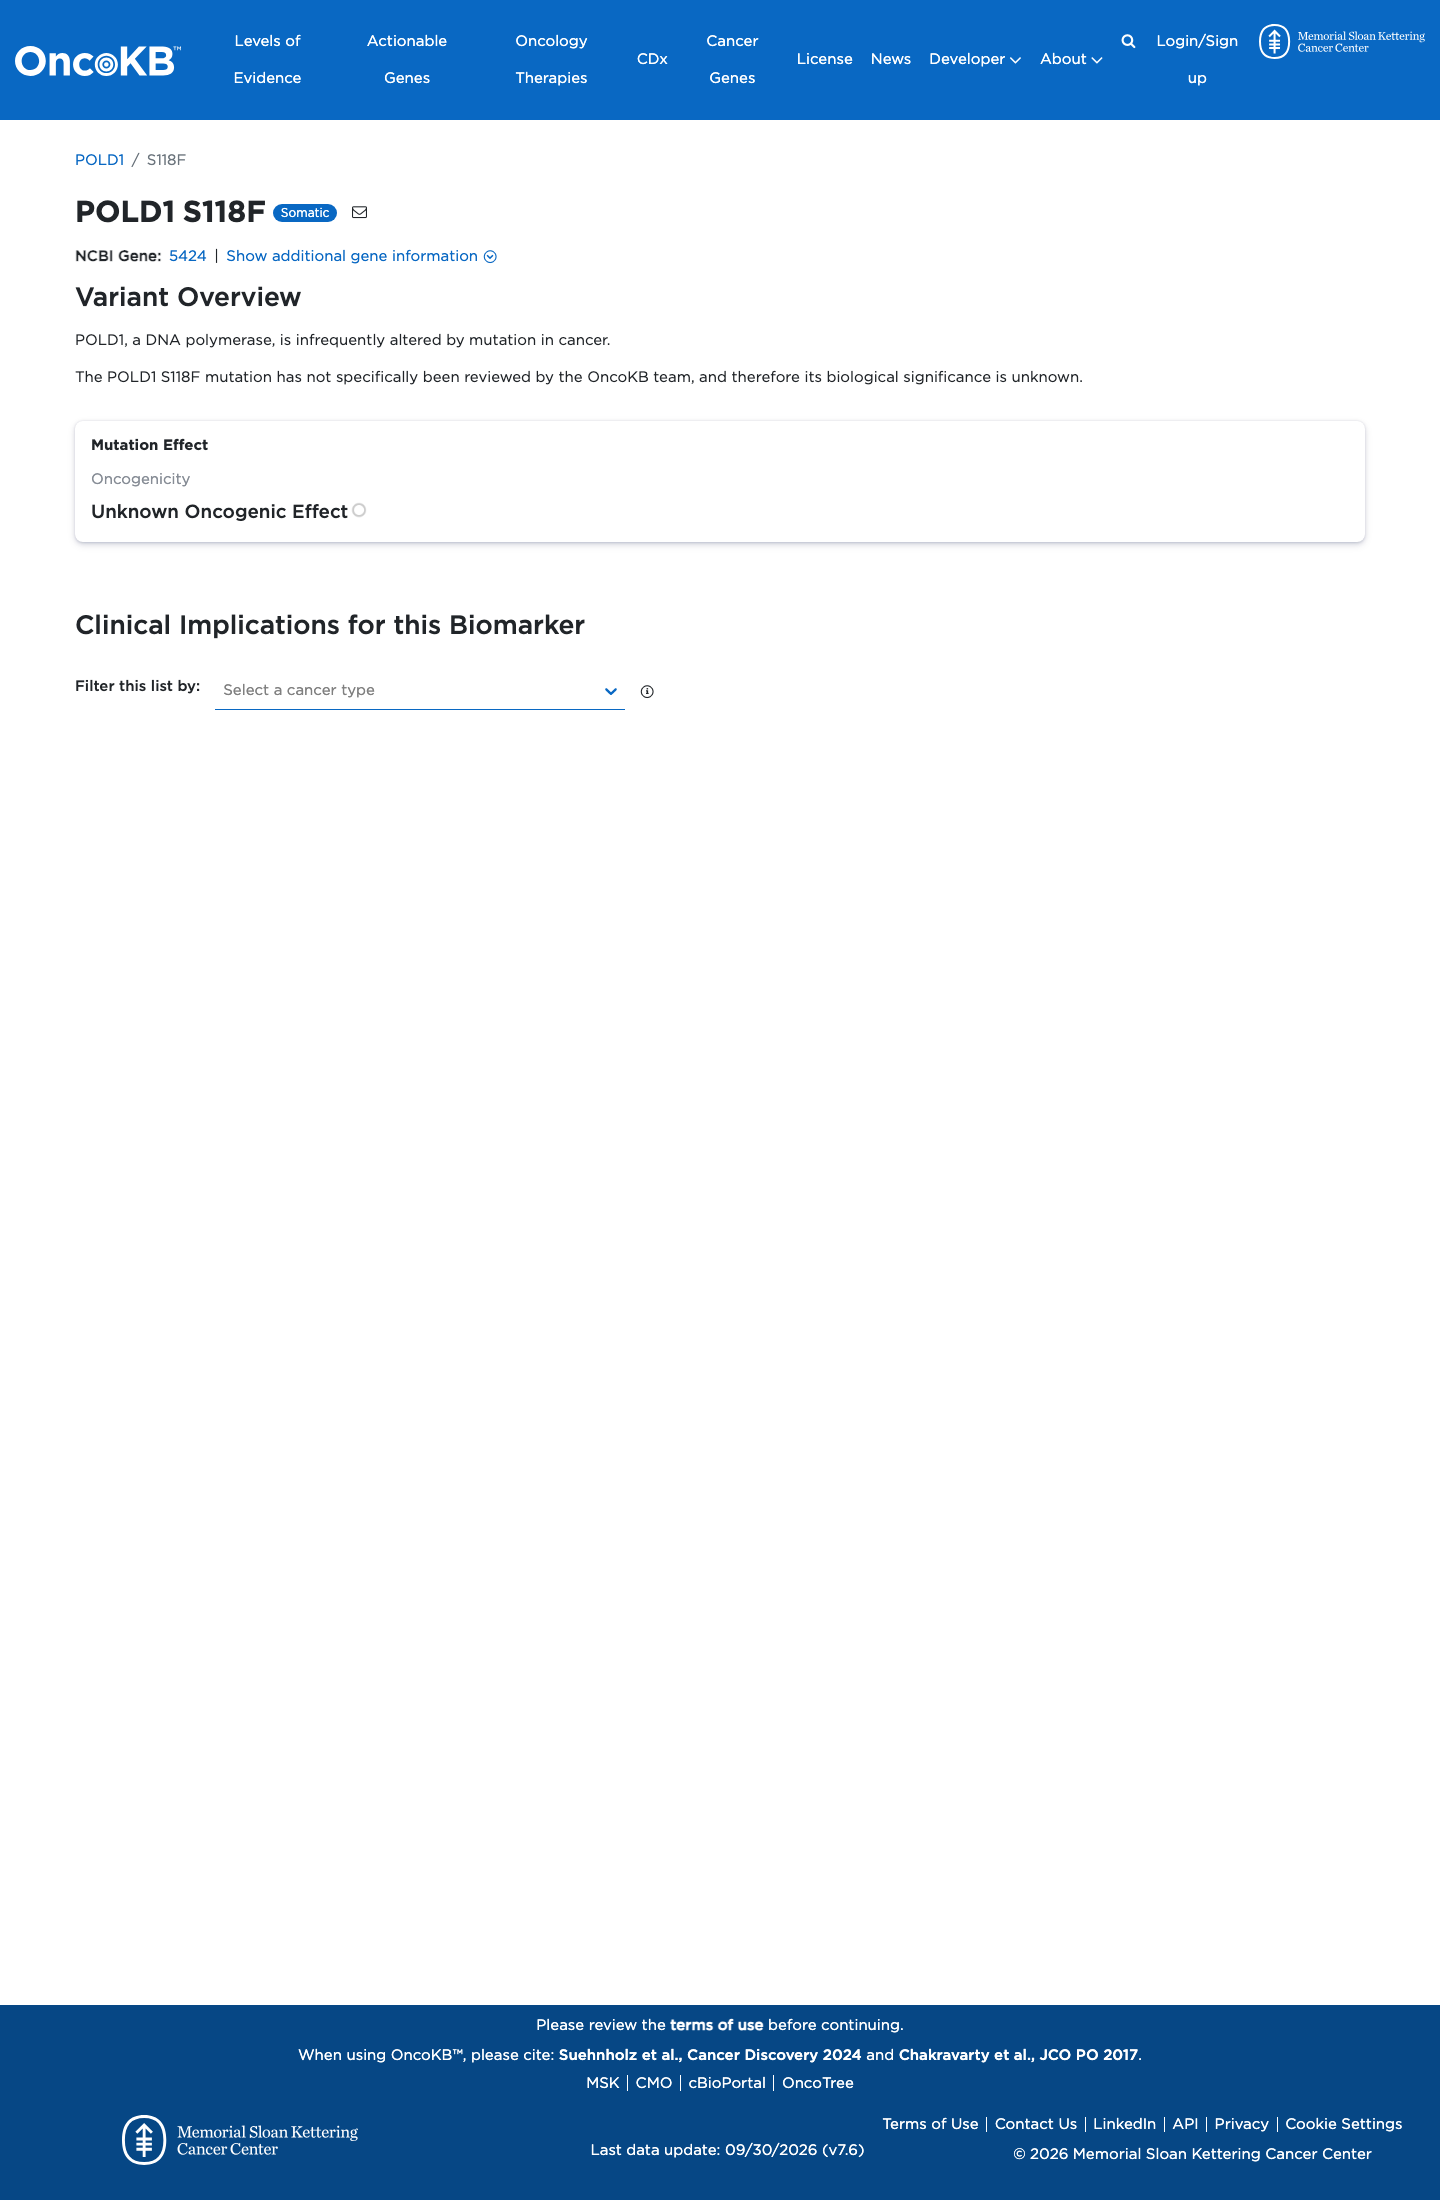
Functional / OncoKB screenshot
Functional Unknown Oncogenic Effect
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. POLD1, a DNA polymerase, is infrequently altered by mutation in cancer.
OncoKB ↗
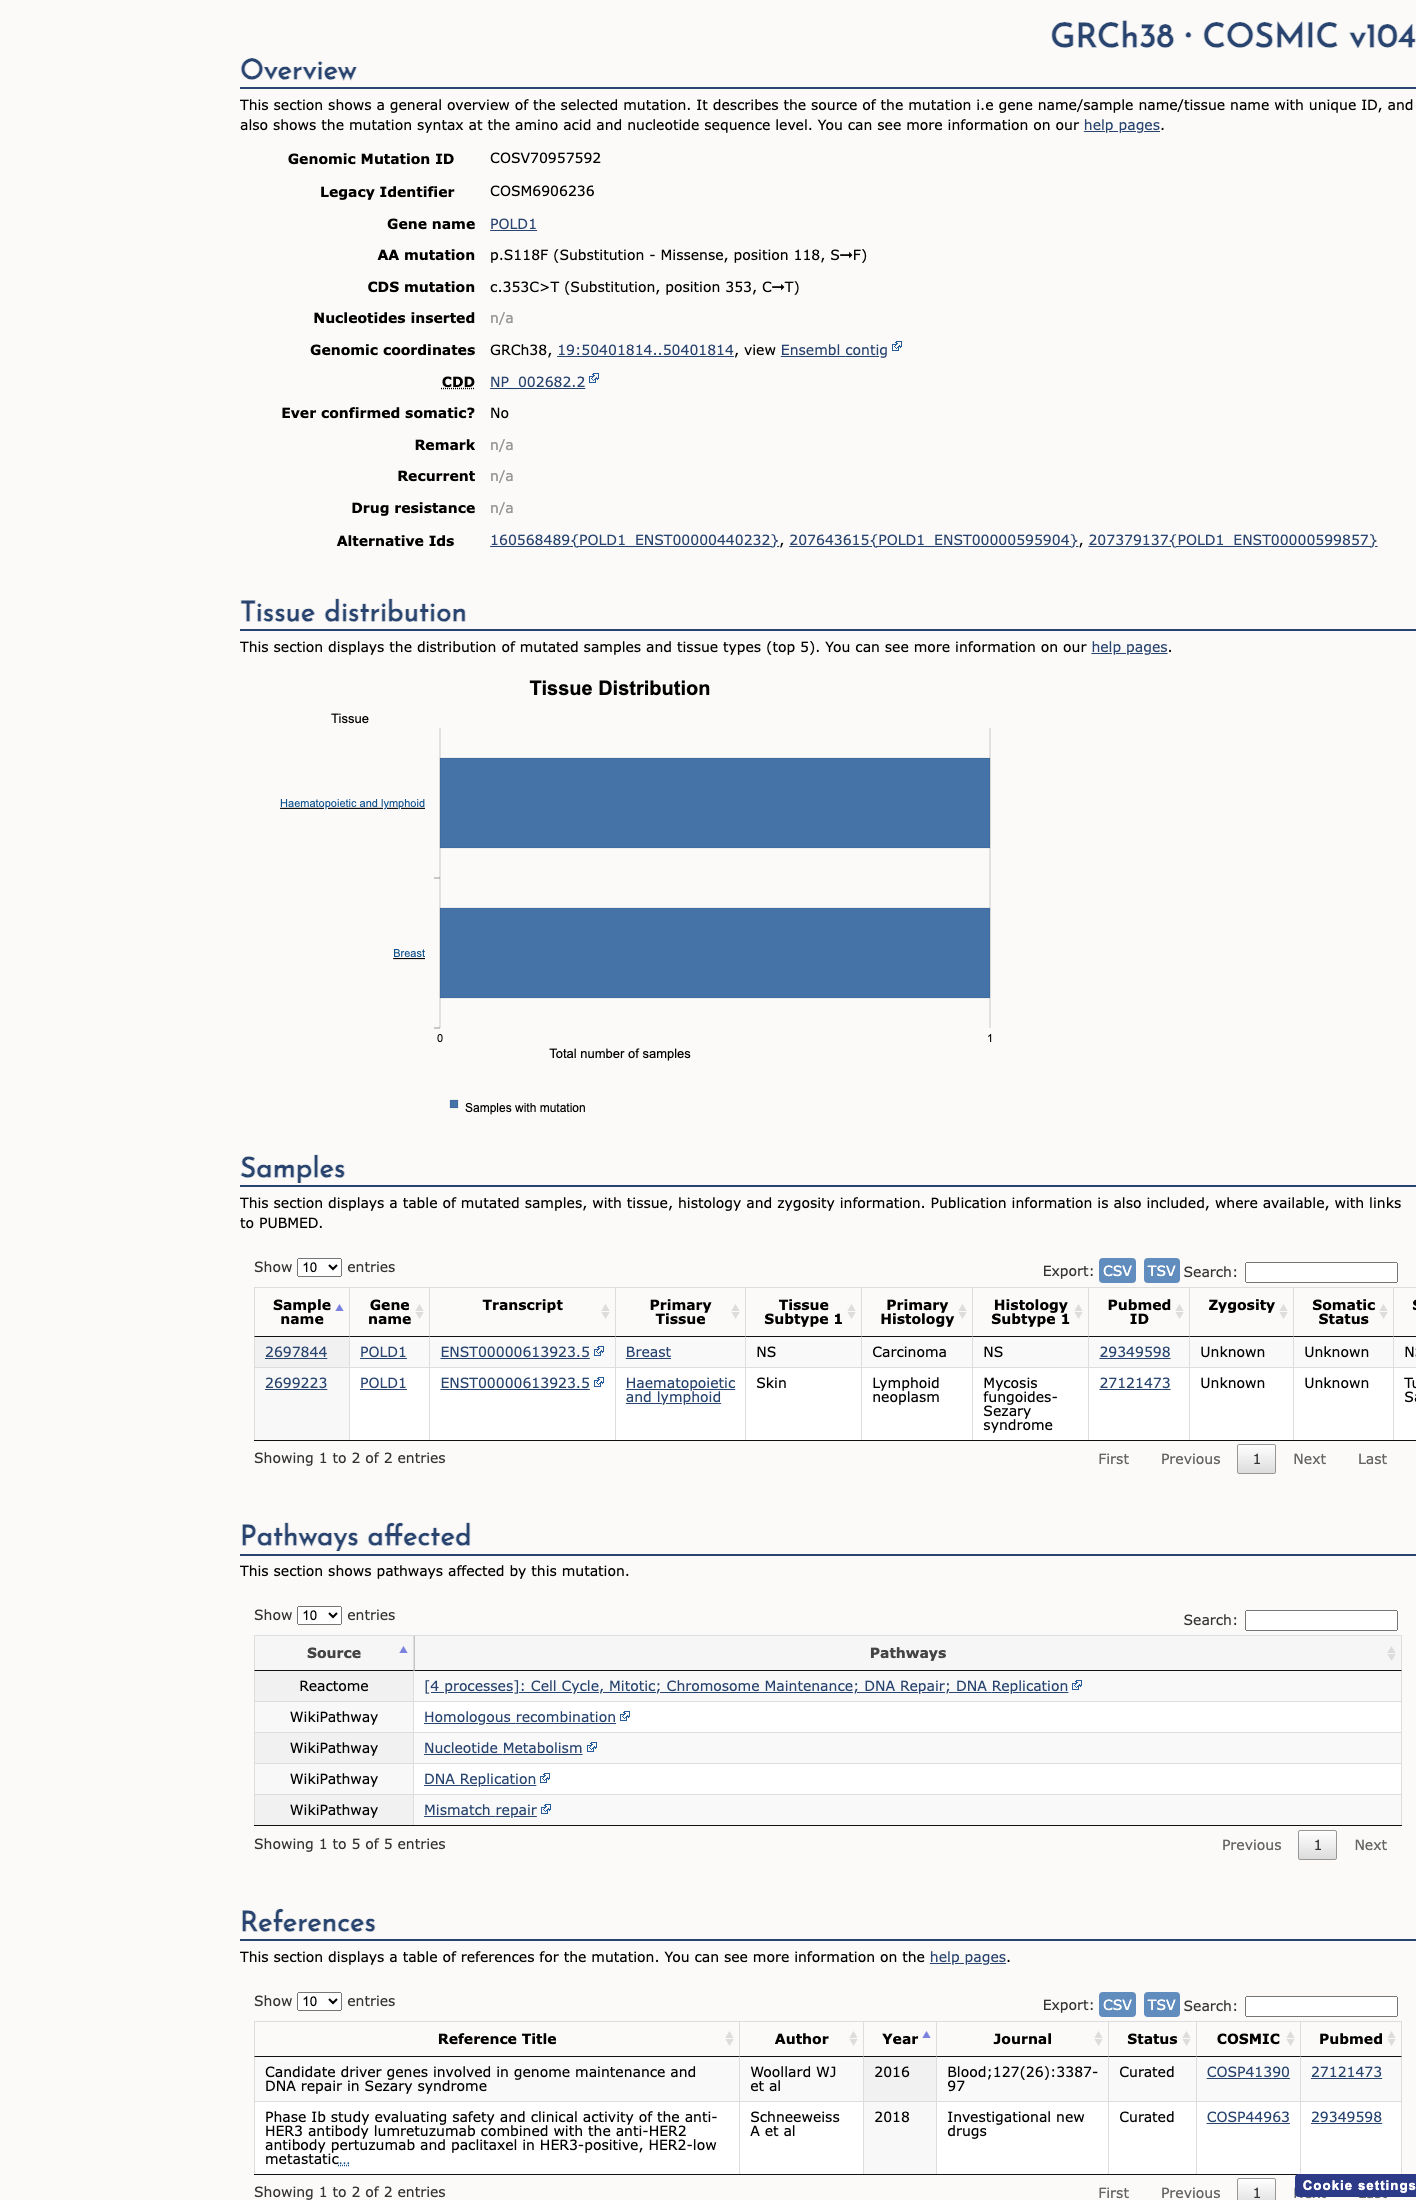
COSMIC screenshot
COSMIC
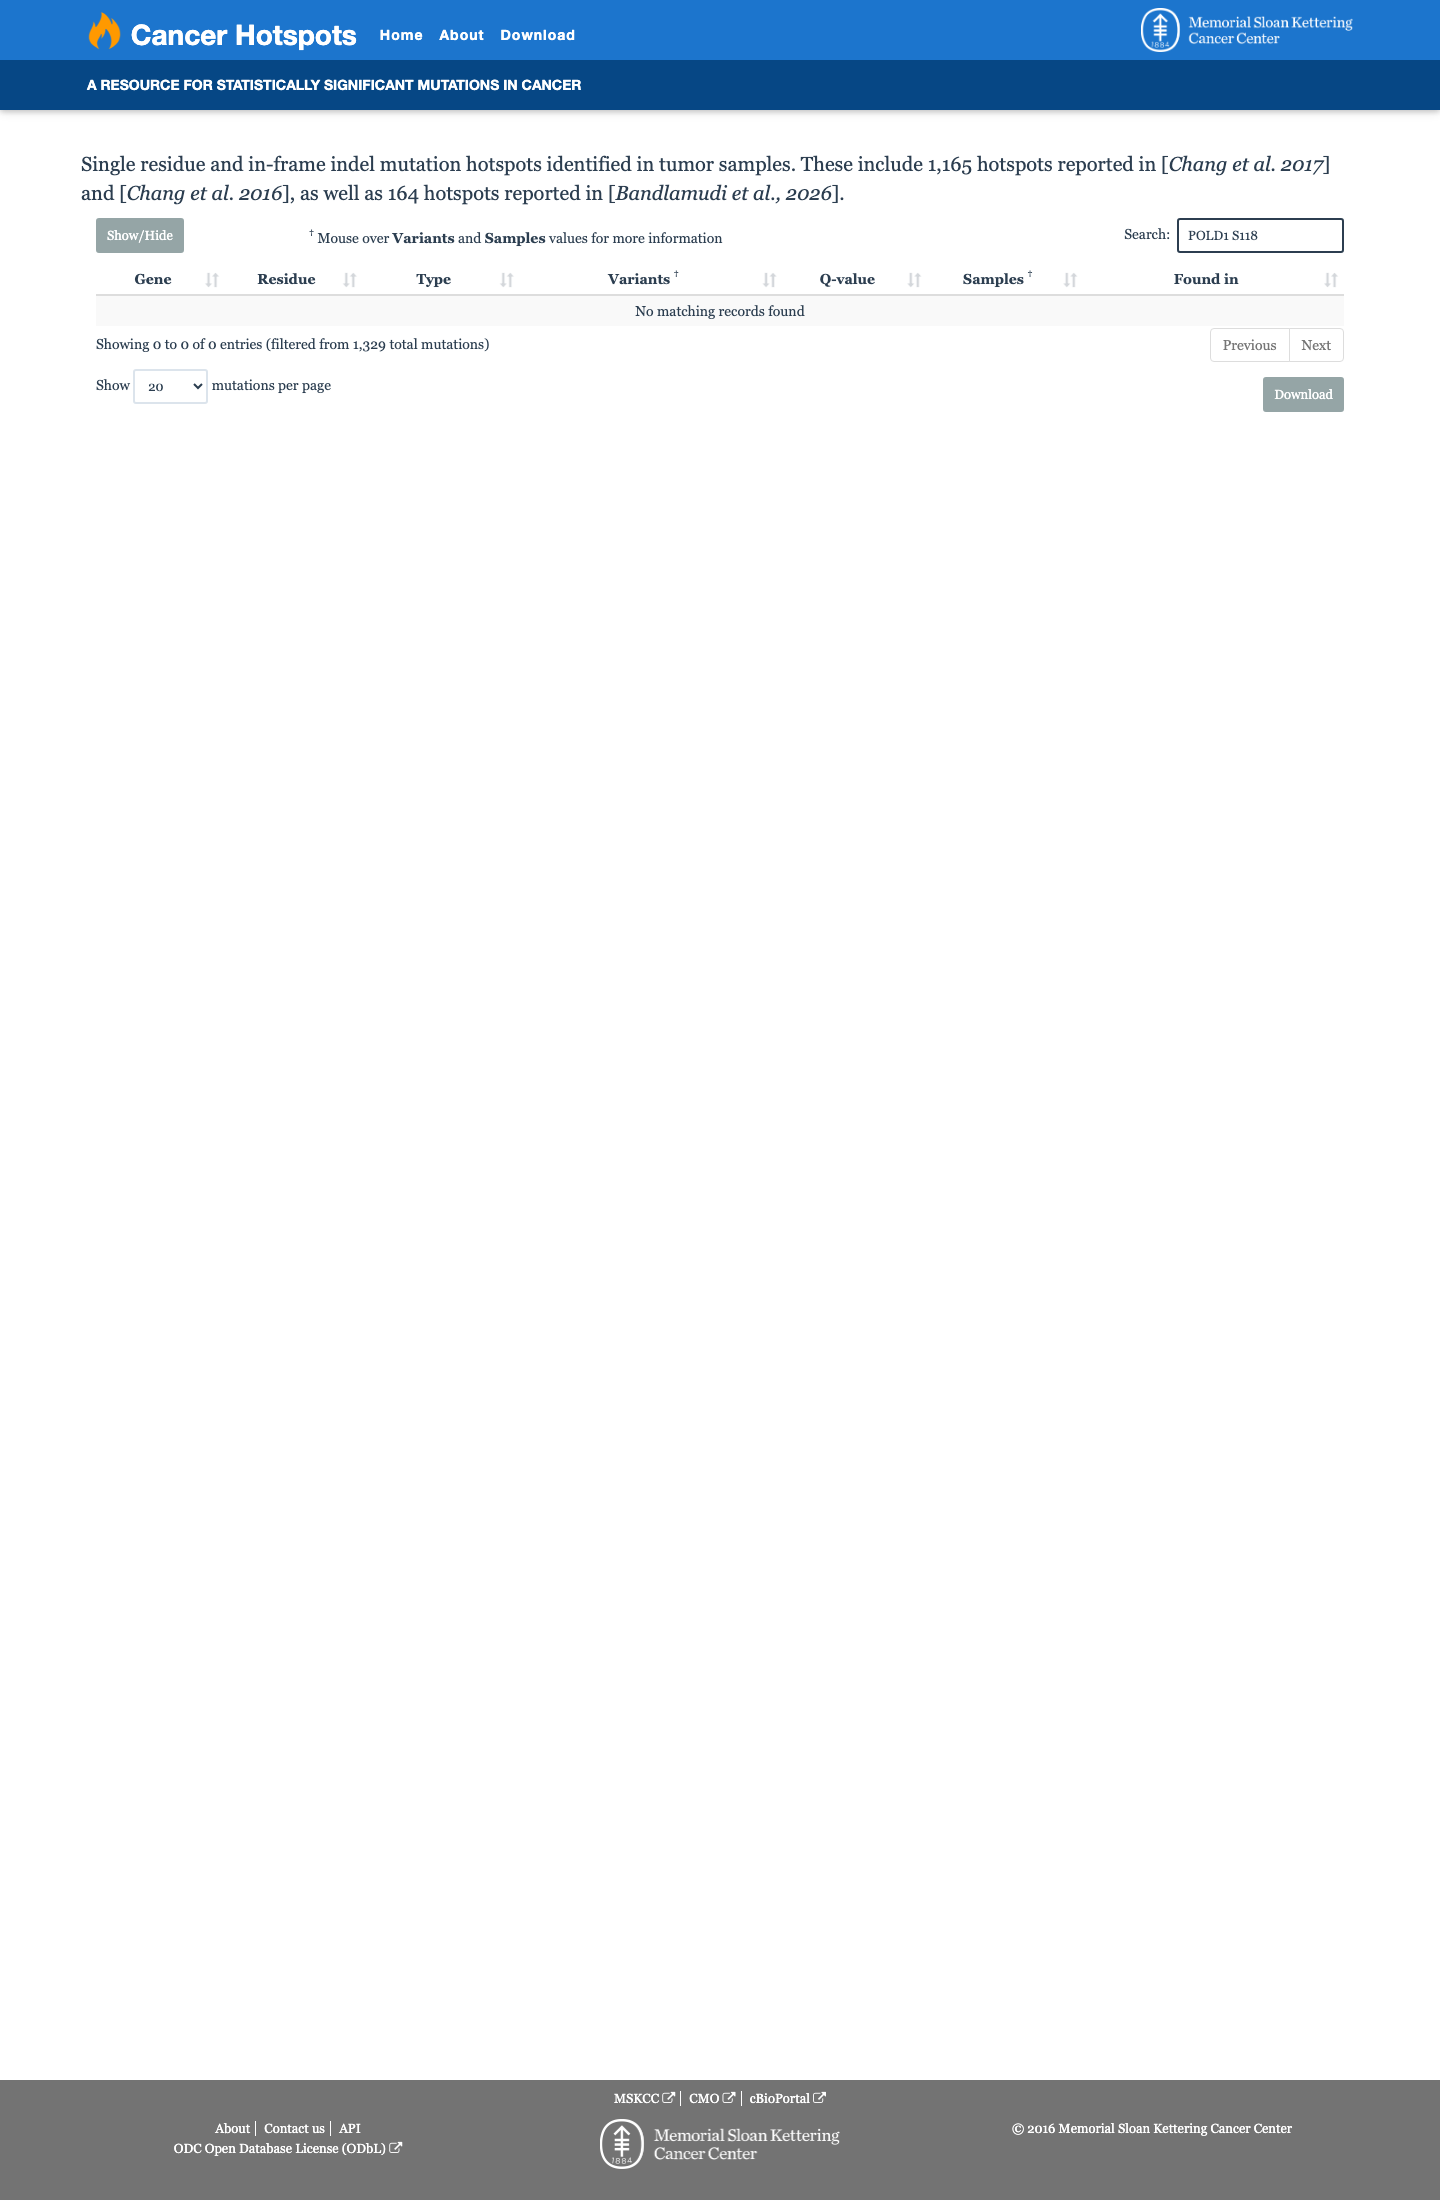
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has previously been reported in somatic cancers (COSMIC; COSV70957592, n = 2 times).
Hotspots
This variant does not lie in a statistically significant hotspot.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 3 PMIDs not cited in assessment
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology. CLINVAR
26467025 ↗ A Standardized DNA Variant Scoring System for Pathogenicity Assessments in Mendelian Disorders. CLINVAR
25394175 ↗ A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment. CLINVAR