Back
NM_005359.6:c.454+23C>T
p.? · SMAD4
ACMG/AMP
0%
complete
Final classification
VUS
BP4
SMAD4
c.454+23C>T
p.?
unknown · exon 4i

SMAD4 encodes a protein that helps transmit TGF-beta and bone morphogenetic protein signals to the nucleus, where it controls genes involved in cell growth and tissue development. It acts as a tumor suppressor and is associated with juvenile polyposis syndrome and hereditary hemorrhagic telangiectasia. Loss or alteration of SMAD4 is especially linked to pancreatic cancer and can also occur in colorectal and lung cancers.

This variant

SMAD4 encodes a tumor-suppressor signaling mediator whose disruption is associated with juvenile polyposis syndrome, hereditary hemorrhagic telangiectasia, and several cancers.

Transcript
NM_005359.6
HGVS · transcript:coding
NM_005359.6:c.454+23C>T
GRCh38
chr18:51049347 C>T
GRCh37
chr18:48575717 C>T
VUS: BP4 (supporting) was the only applied criterion, and no generic ACMG/AMP combination threshold for a definitive classification was met.
Classification rationale
BP4 VUS
SMAD4 c.454+23C>T unknown · exon 4i

VUS: BP4 (supporting) was met because SpliceAI maximum delta 0.008 is below the <=0.1 threshold for no significant predicted splice impact.

BP4 VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_005359.6 · variants mapped to exon structure
SMAD4 NM_005359.6
Fetching transcript structure from UCSC…
Applied criteria · 1 applied · 18 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 1
Strength Supporting Moderate Strong Very strong
BP4 supporting Benign
Met, supporting: SpliceAI maximum delta 0.008 is below the <=0.1 BP4 threshold.
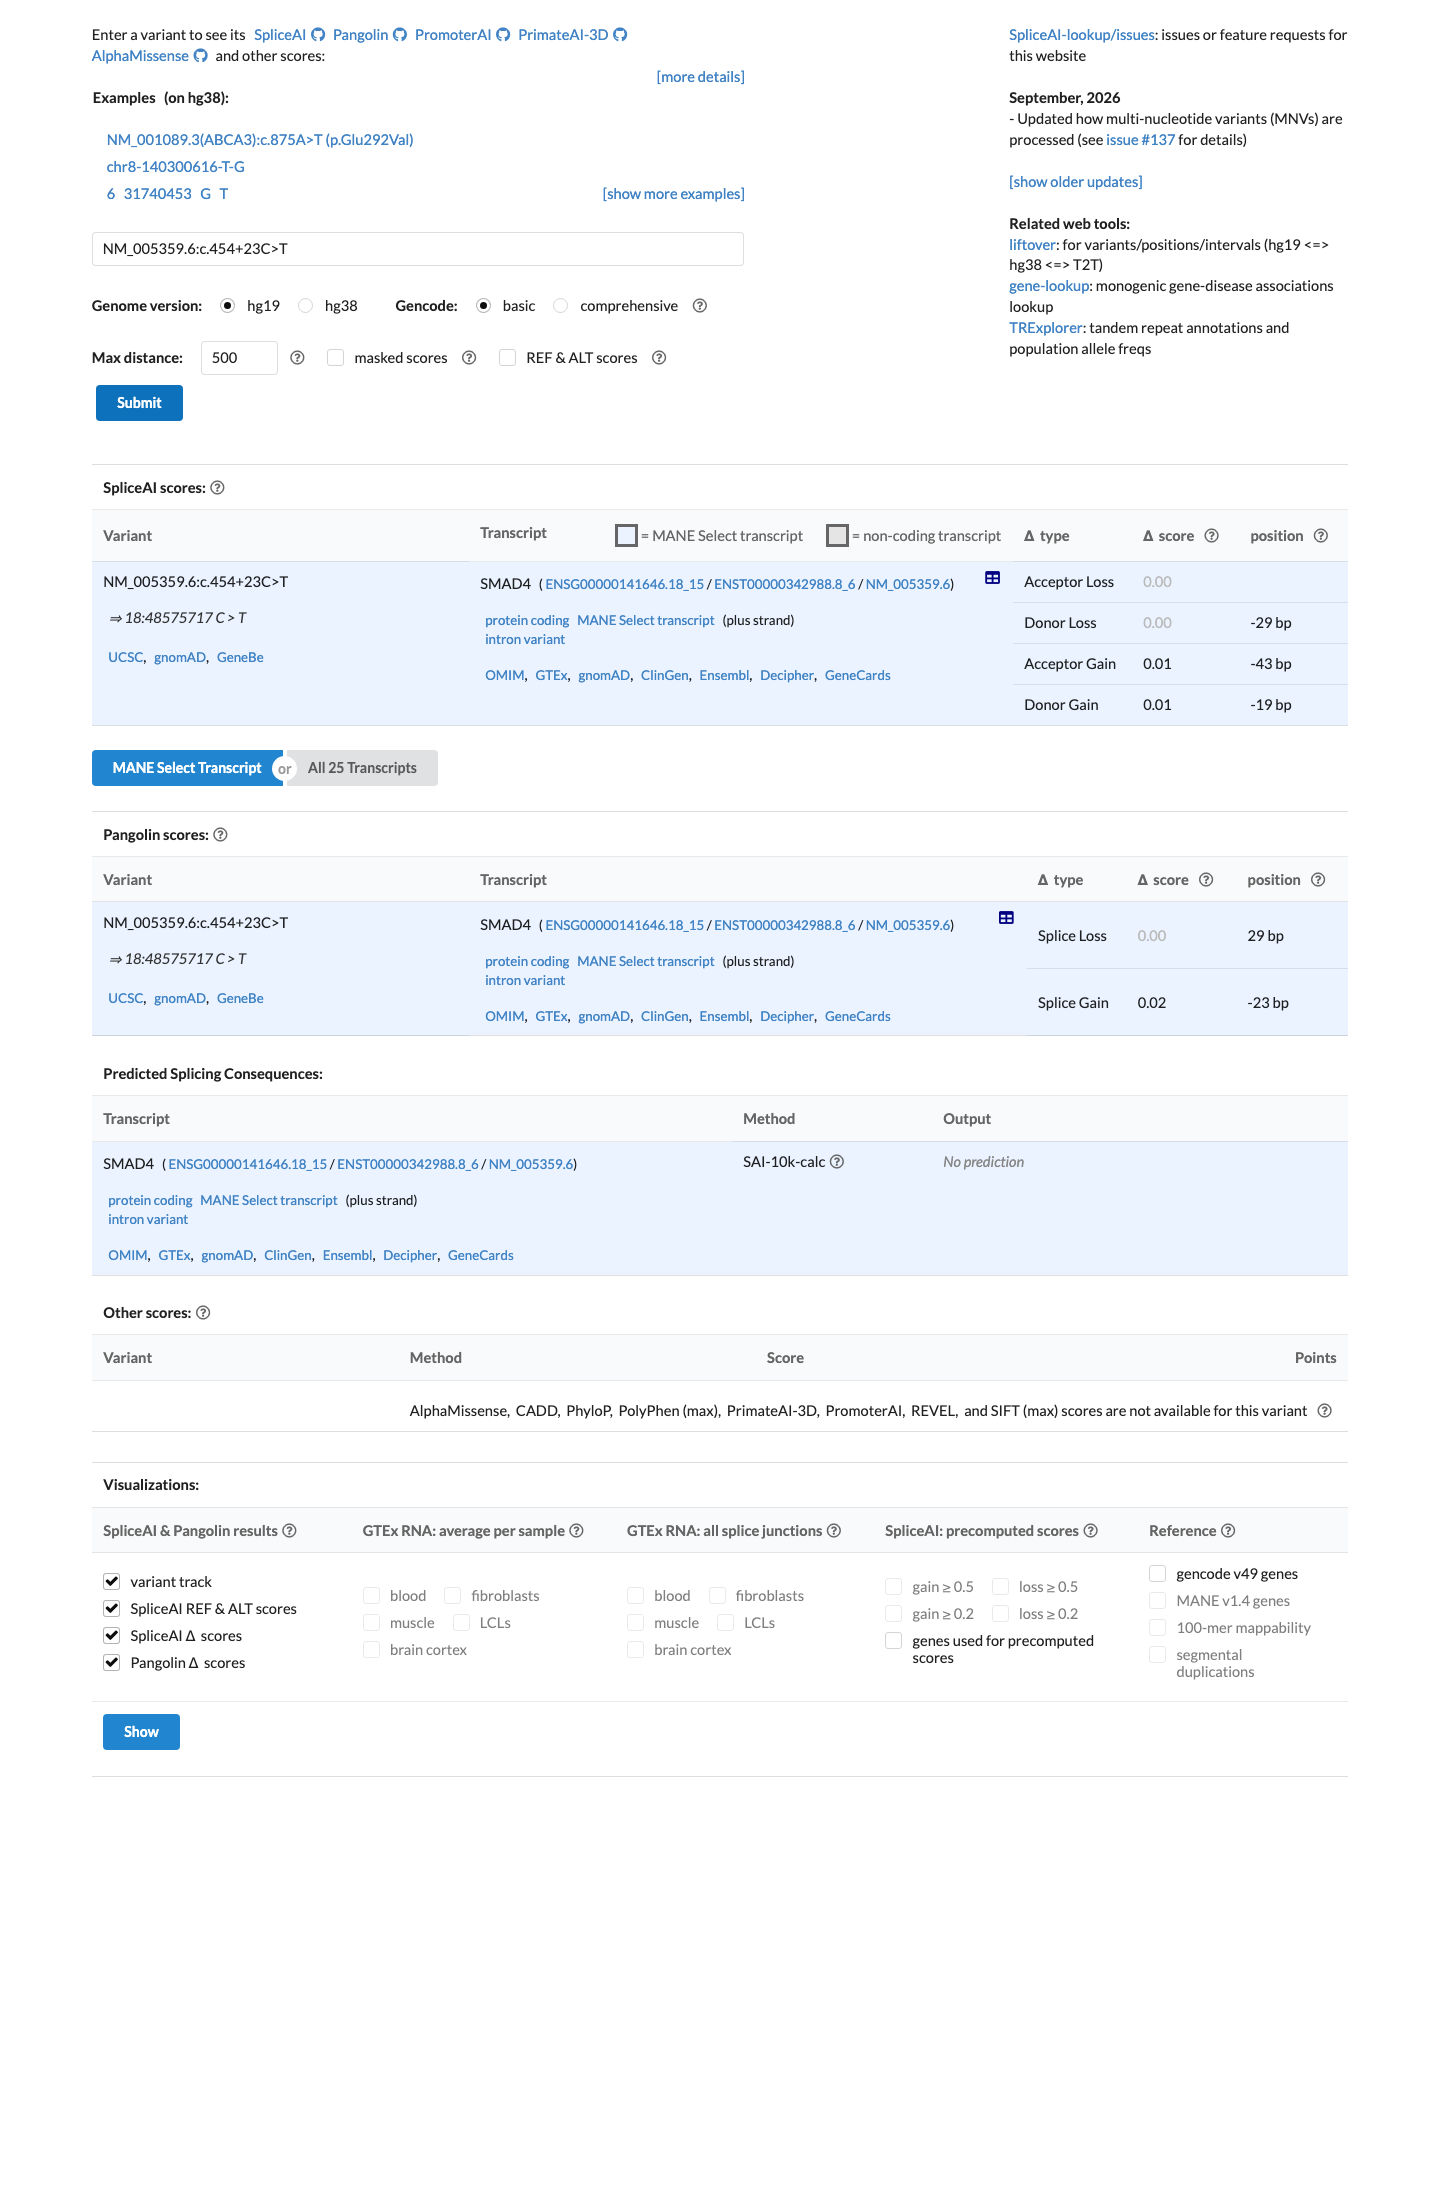
The variant is intronic at c.454+23 and is therefore within BP4 splice-impact scope.SpliceAI maximum delta score is 0.008 (DS_AG 0.008, DS_DG 0.007, DS_AL 0.0, DS_DL 0.0).The generic supporting BP4 threshold is <=0.1, from the Jaganathan et al. 2019 SpliceAI calibration (PMID:30661751); 0.008 meets it.
Assessed · not applied · 6 not met · 12 not assessed
Pathogenic
PS2 Not assessed: no documented parental testing or confirmed de novo occurrence is available for this variant.
PS3 Not assessed: no variant-specific validated functional assay or assay result is available for NM_005359.6:c.454+23C>T.
PS4 Not assessed: no case-control cohort, odds ratio, or exact-variant prevalence comparison is available to establish affected-individual enrichment.
PM2 Not met: gnomAD v4.1 overall AF 0.000302921 exceeds the generic PM2 threshold of 0.0001.
PM3 Not assessed: no affected-proband observation, second pathogenic variant, phase result, or inheritance information is documented for this intronic SMAD4 variant.
PM6 Not assessed: no affected proband with a presumed de novo variant is documented, even without parental confirmation.
PP1 Not assessed: zero informative affected-relative genotypes or meioses are documented for segregation analysis.
PP3 Not met: SpliceAI maximum delta 0.008 is below the >=0.2 supporting PP3 threshold.
PP4 Not assessed: no patient phenotype, clinical diagnosis, or disease-specific family history is available to establish a highly specific phenotype match.
PP5 Not met: the exact ClinVar record has zero expert-panel submissions and only single-submitter Likely benign assertions.
Benign
BA1 Not met: the highest default all-comers gnomAD v4.1 population AF is 0.005251535, below the generic BA1 threshold of 0.05.
BS1 Not met: the highest default all-comers gnomAD v4.1 population AF is 0.005251535, below the generic BS1 threshold of 0.01.
BS2 Not assessed: one gnomAD homozygote is reported, but age, phenotype, and clinical health of that individual are unavailable.
BS3 Not assessed: no variant-specific validated functional assay or assay result is available to demonstrate preserved function for NM_005359.6:c.454+23C>T.
BS4 Not assessed: no affected-family testing or documented non-segregation observation is available for this variant.
BP2 Not assessed: no additional pathogenic variant or validated cis/trans phase result is documented for this intronic SMAD4 variant.
BP5 Not assessed: no affected-individual phenotype or confirmed alternate molecular diagnosis is documented for this exact variant.
BP6 Not met: the exact ClinVar record has zero expert-panel submissions and only single-submitter Likely benign assertions.
N/A · 9 PVS1 · PS1 · PM1 · PM4 · PM5 · PP2 · BP1 · BP3 · BP7
Research & evidence
Population frequency
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
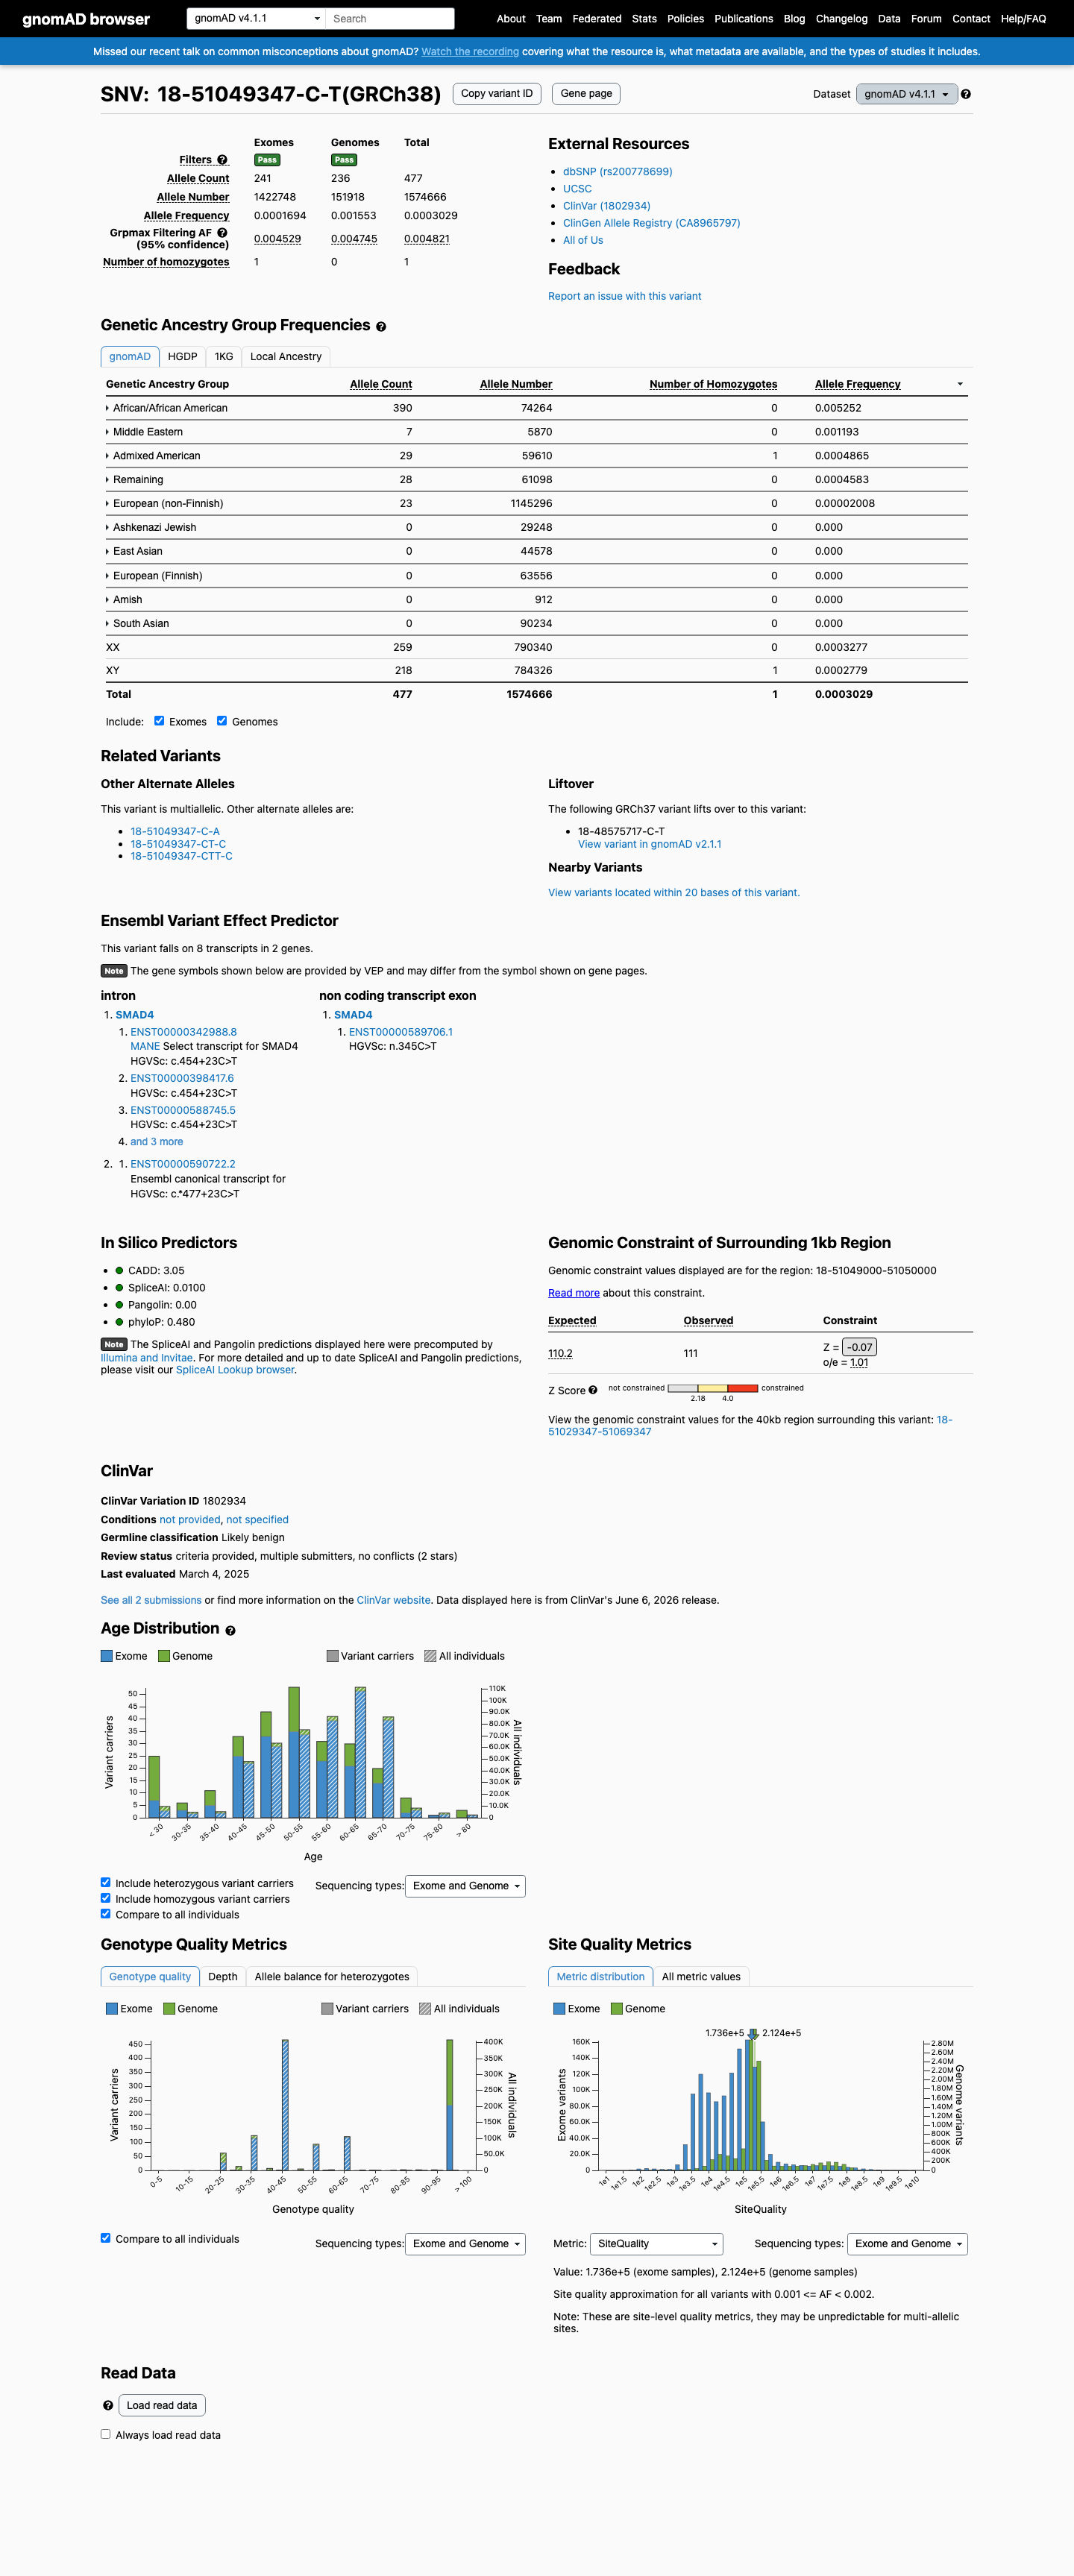
This variant is present in gnomAD v4.1 (AF= 0.000302921; MAF= 0.03029%, 477/1574666 alleles, homozygotes = 1) and has highest observed frequency in the African/African American population (AF= 0.00525154; MAF= 0.52515%, 390/74264 alleles, homozygotes = 0); grpmax FAF= 0.004821.
v2.1
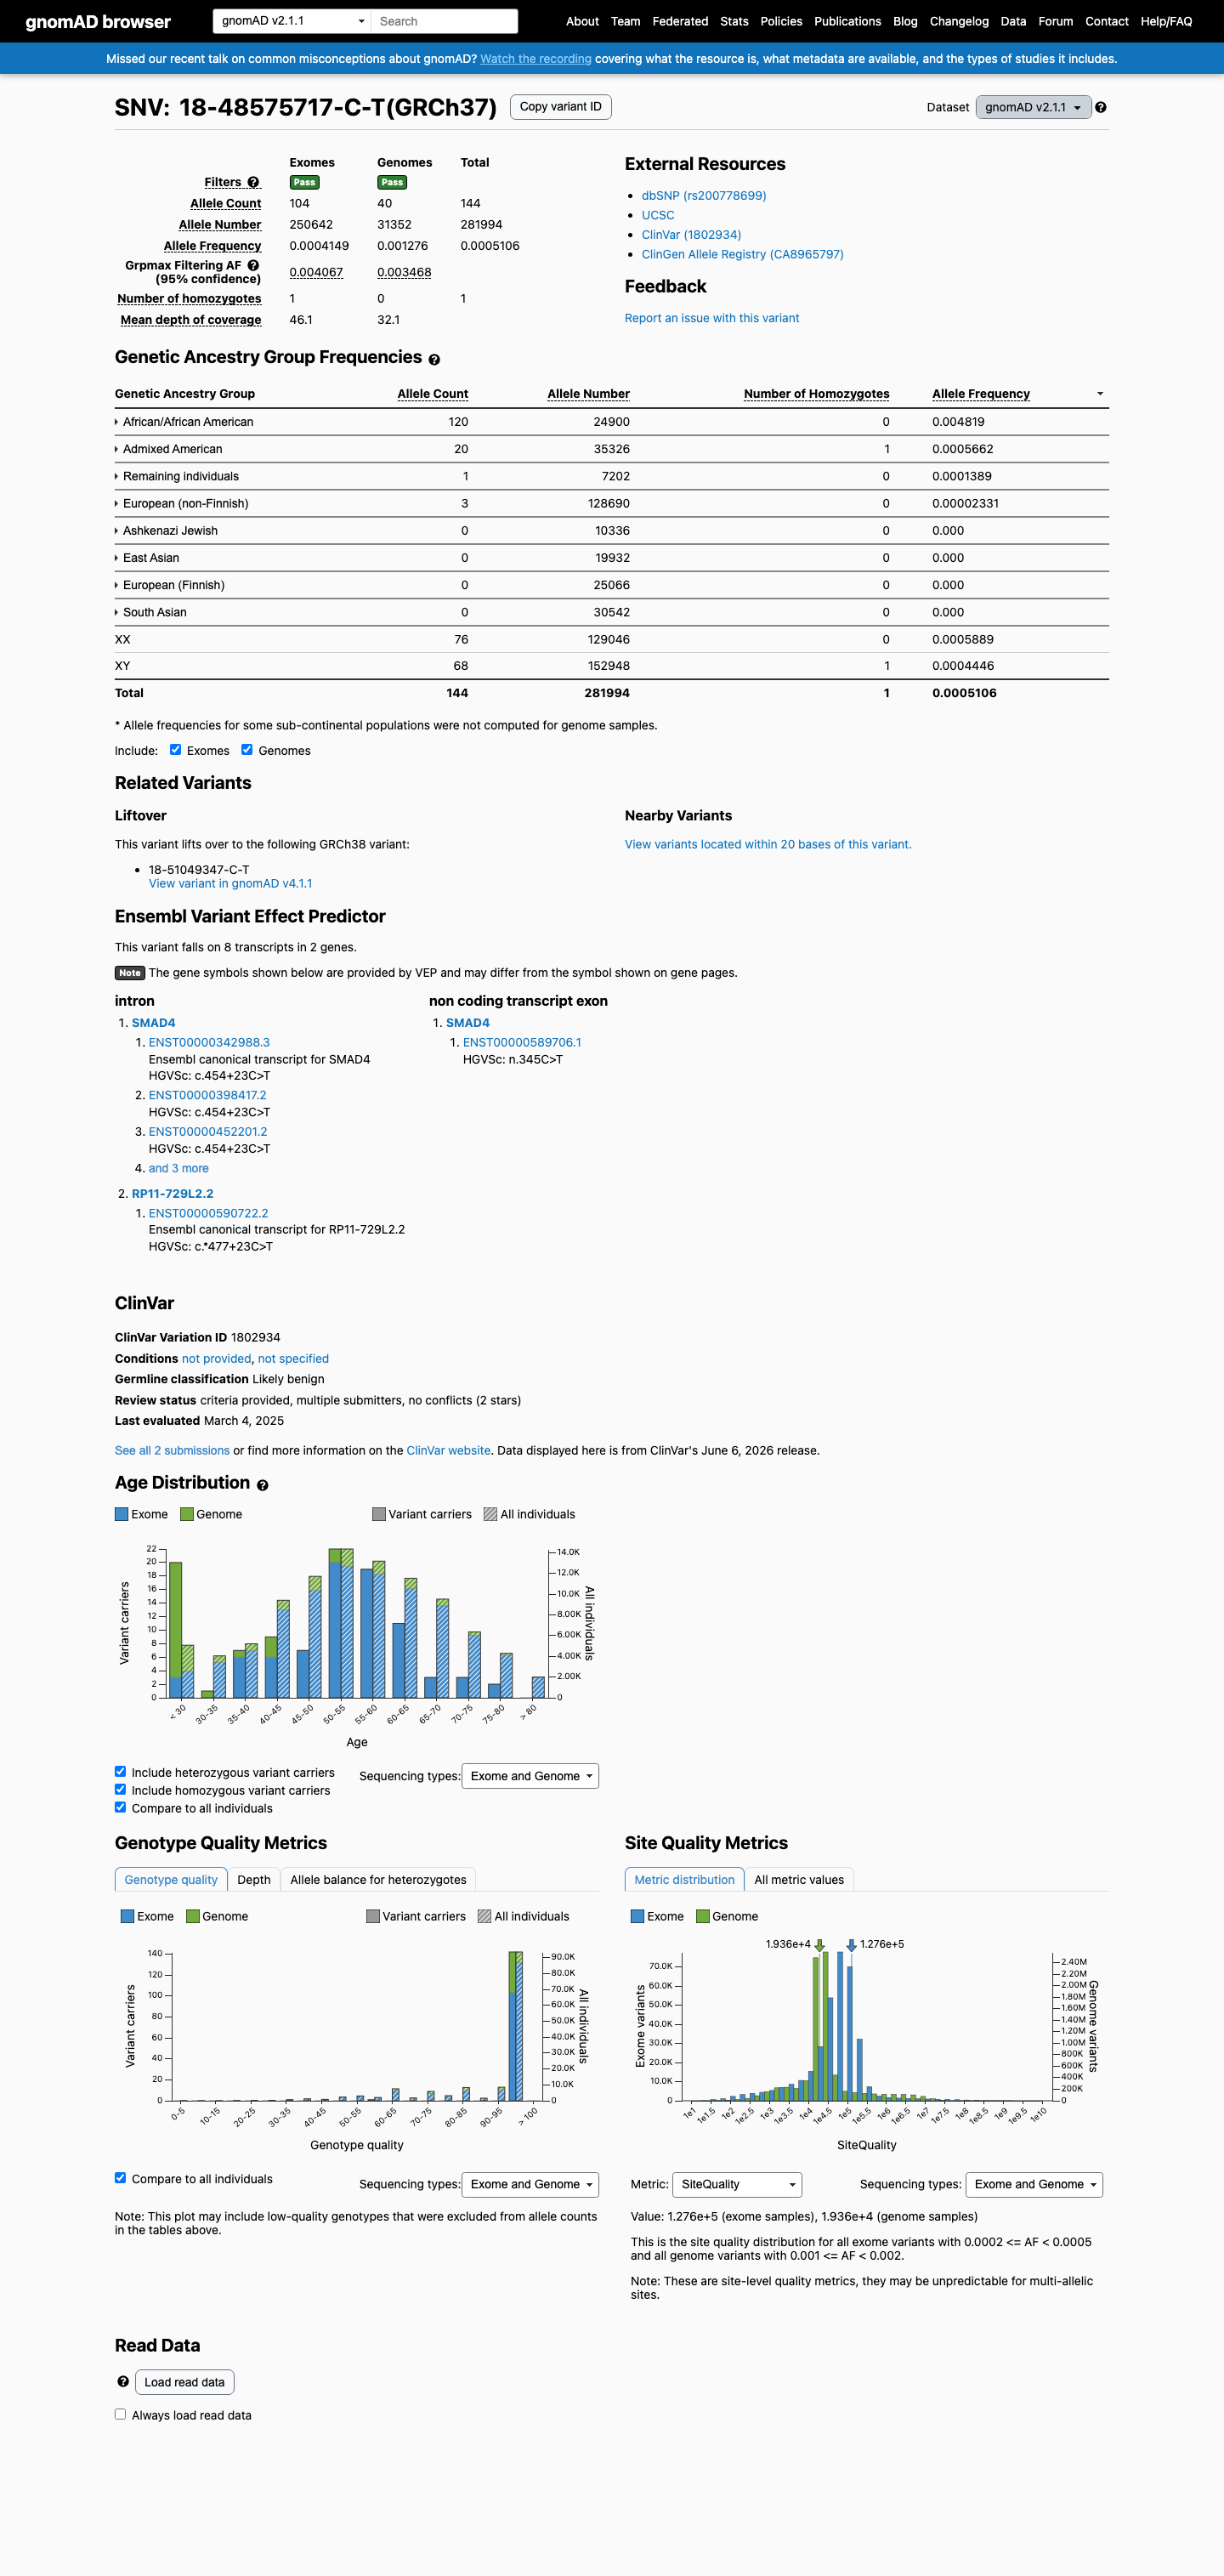
This variant is present in gnomAD v2.1 (AF= 0.000510649; MAF= 0.05106%, 144/281994 alleles, homozygotes = 1) and has highest observed frequency in the African/African American population (AF= 0.00481928; MAF= 0.48193%, 120/24900 alleles, homozygotes = 0); grpmax FAF= 0.00406717.
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.03% · 477 / 1,574,666
1 hom · FAF 0.48%
African/African American
390 / 74,264
0.53%
Middle Eastern
7 / 5,870
0.12%
Admixed American
29 / 59,610
0.049%
1 hom
Remaining individuals
28 / 61,098
0.046%
European (non-Finnish)
23 / 1,145,296
0.002%
+ 5 not observed (European (Finnish), Amish, East Asian, South Asian, Ashkenazi Jewish)
gnomAD v2.1
0.051% · 144 / 281,994
1 hom · FAF 0.41%
African/African American
120 / 24,900
0.48%
Admixed American
20 / 35,326
0.057%
1 hom
Remaining individuals
1 / 7,202
0.014%
European (non-Finnish)
3 / 128,690
0.0023%
+ 4 not observed (Ashkenazi Jewish, East Asian, European (Finnish), South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
ClinVar screenshot
ClinVar
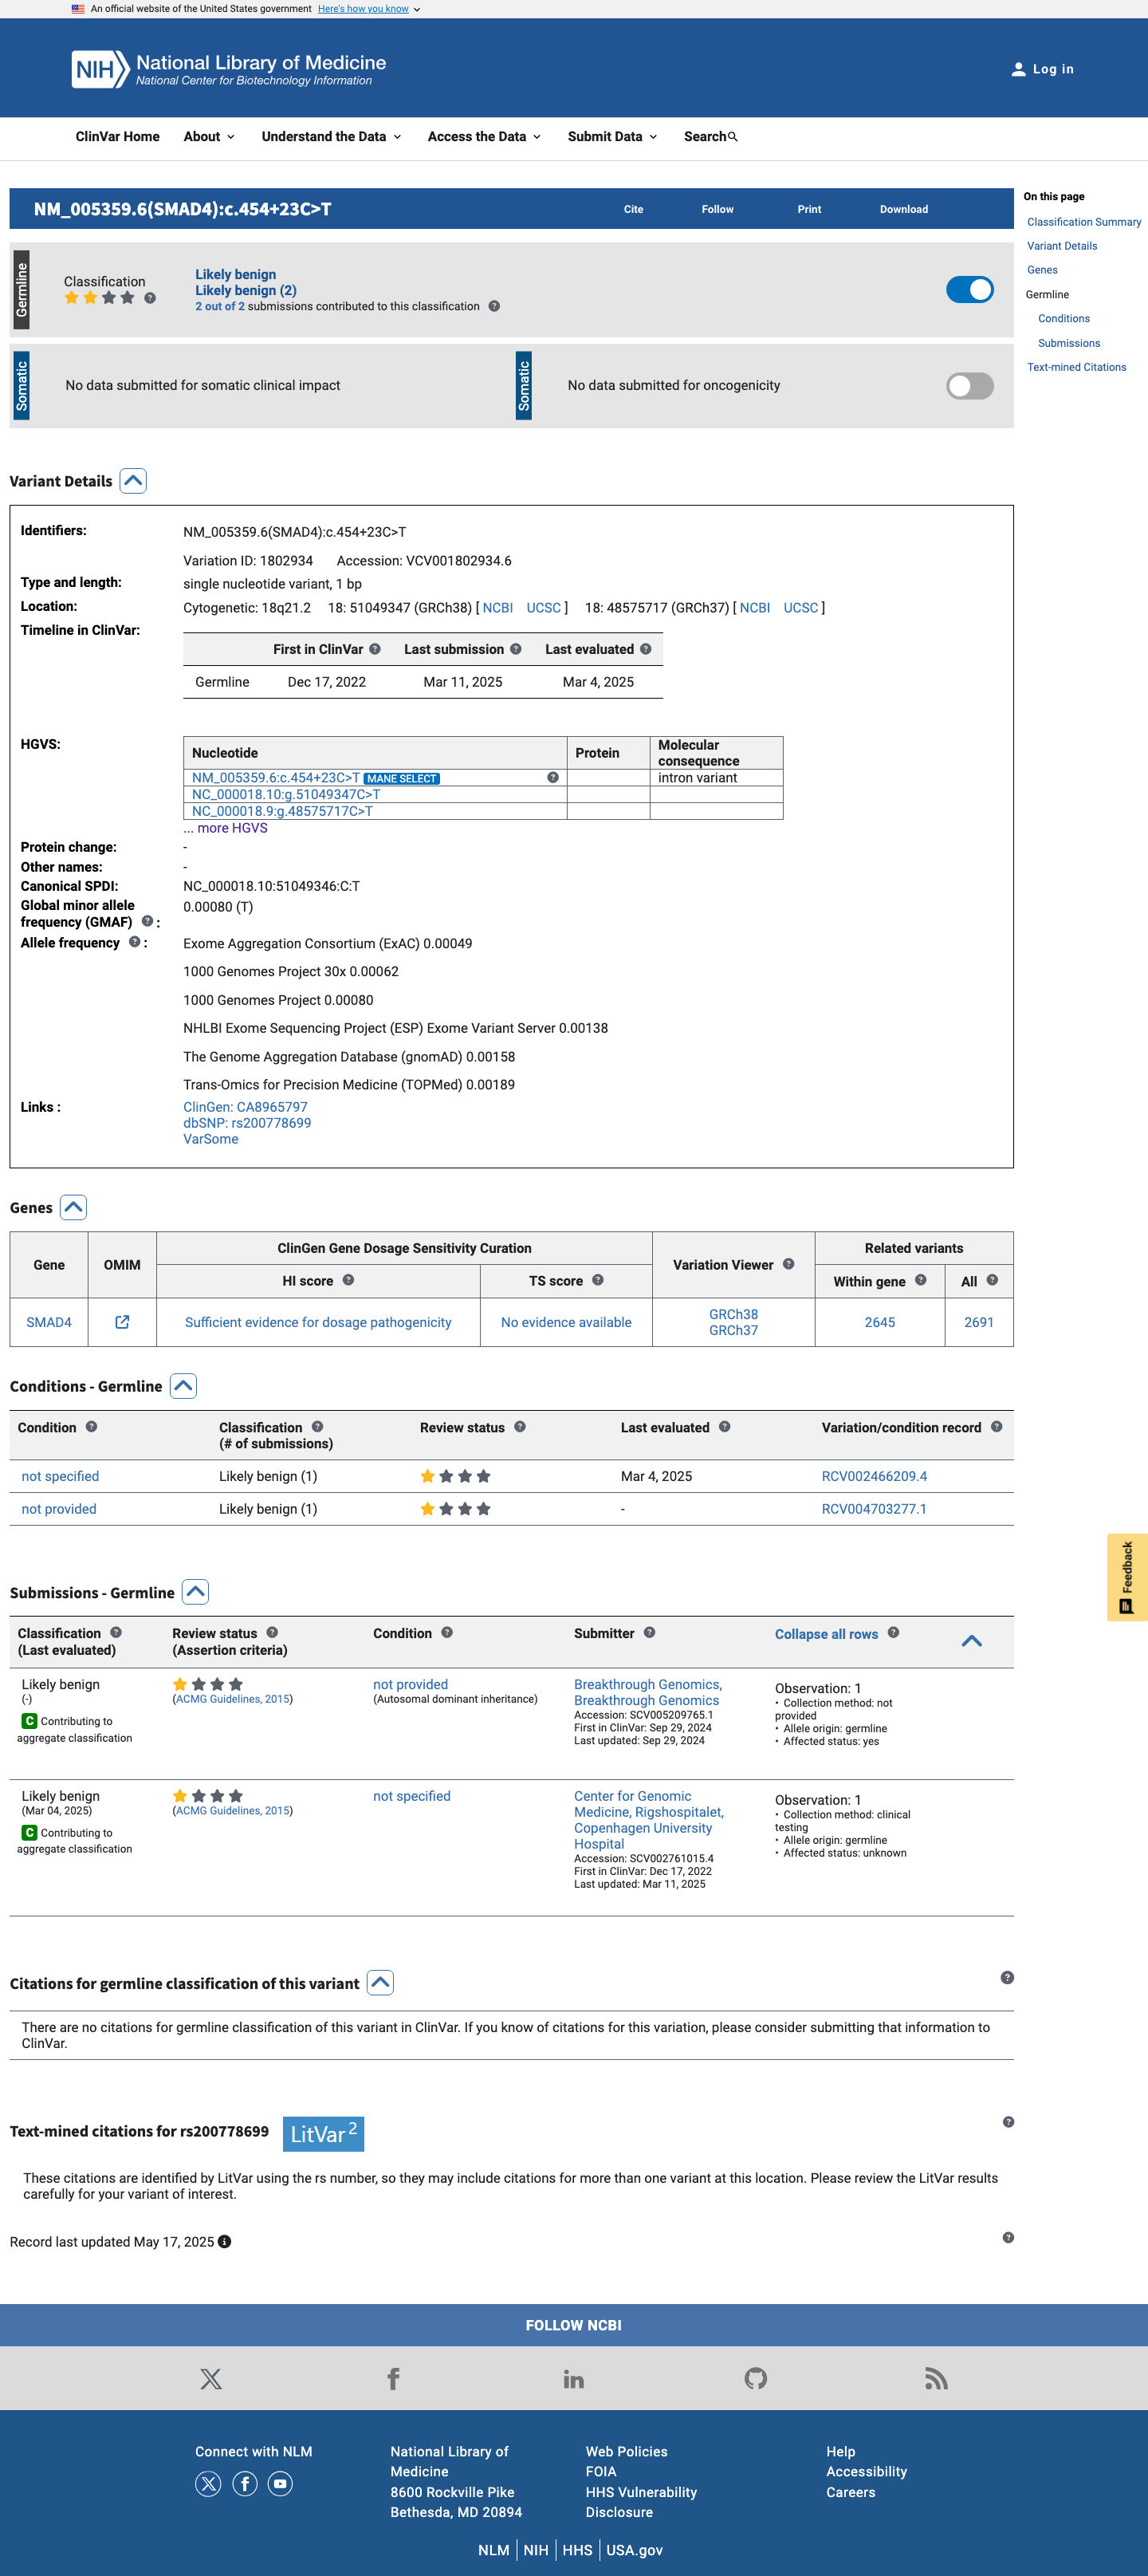
This variant has been reported in ClinVar as Likely benign (1 clinical laboratory). (ClinVarID = 1802934)
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.01).
Functional No data
No calibrated functional assay or RNA evidence was identified for this variant.
OncoKB ↗
COSMIC screenshot
COSMIC
Somatic evidence
COSMIC
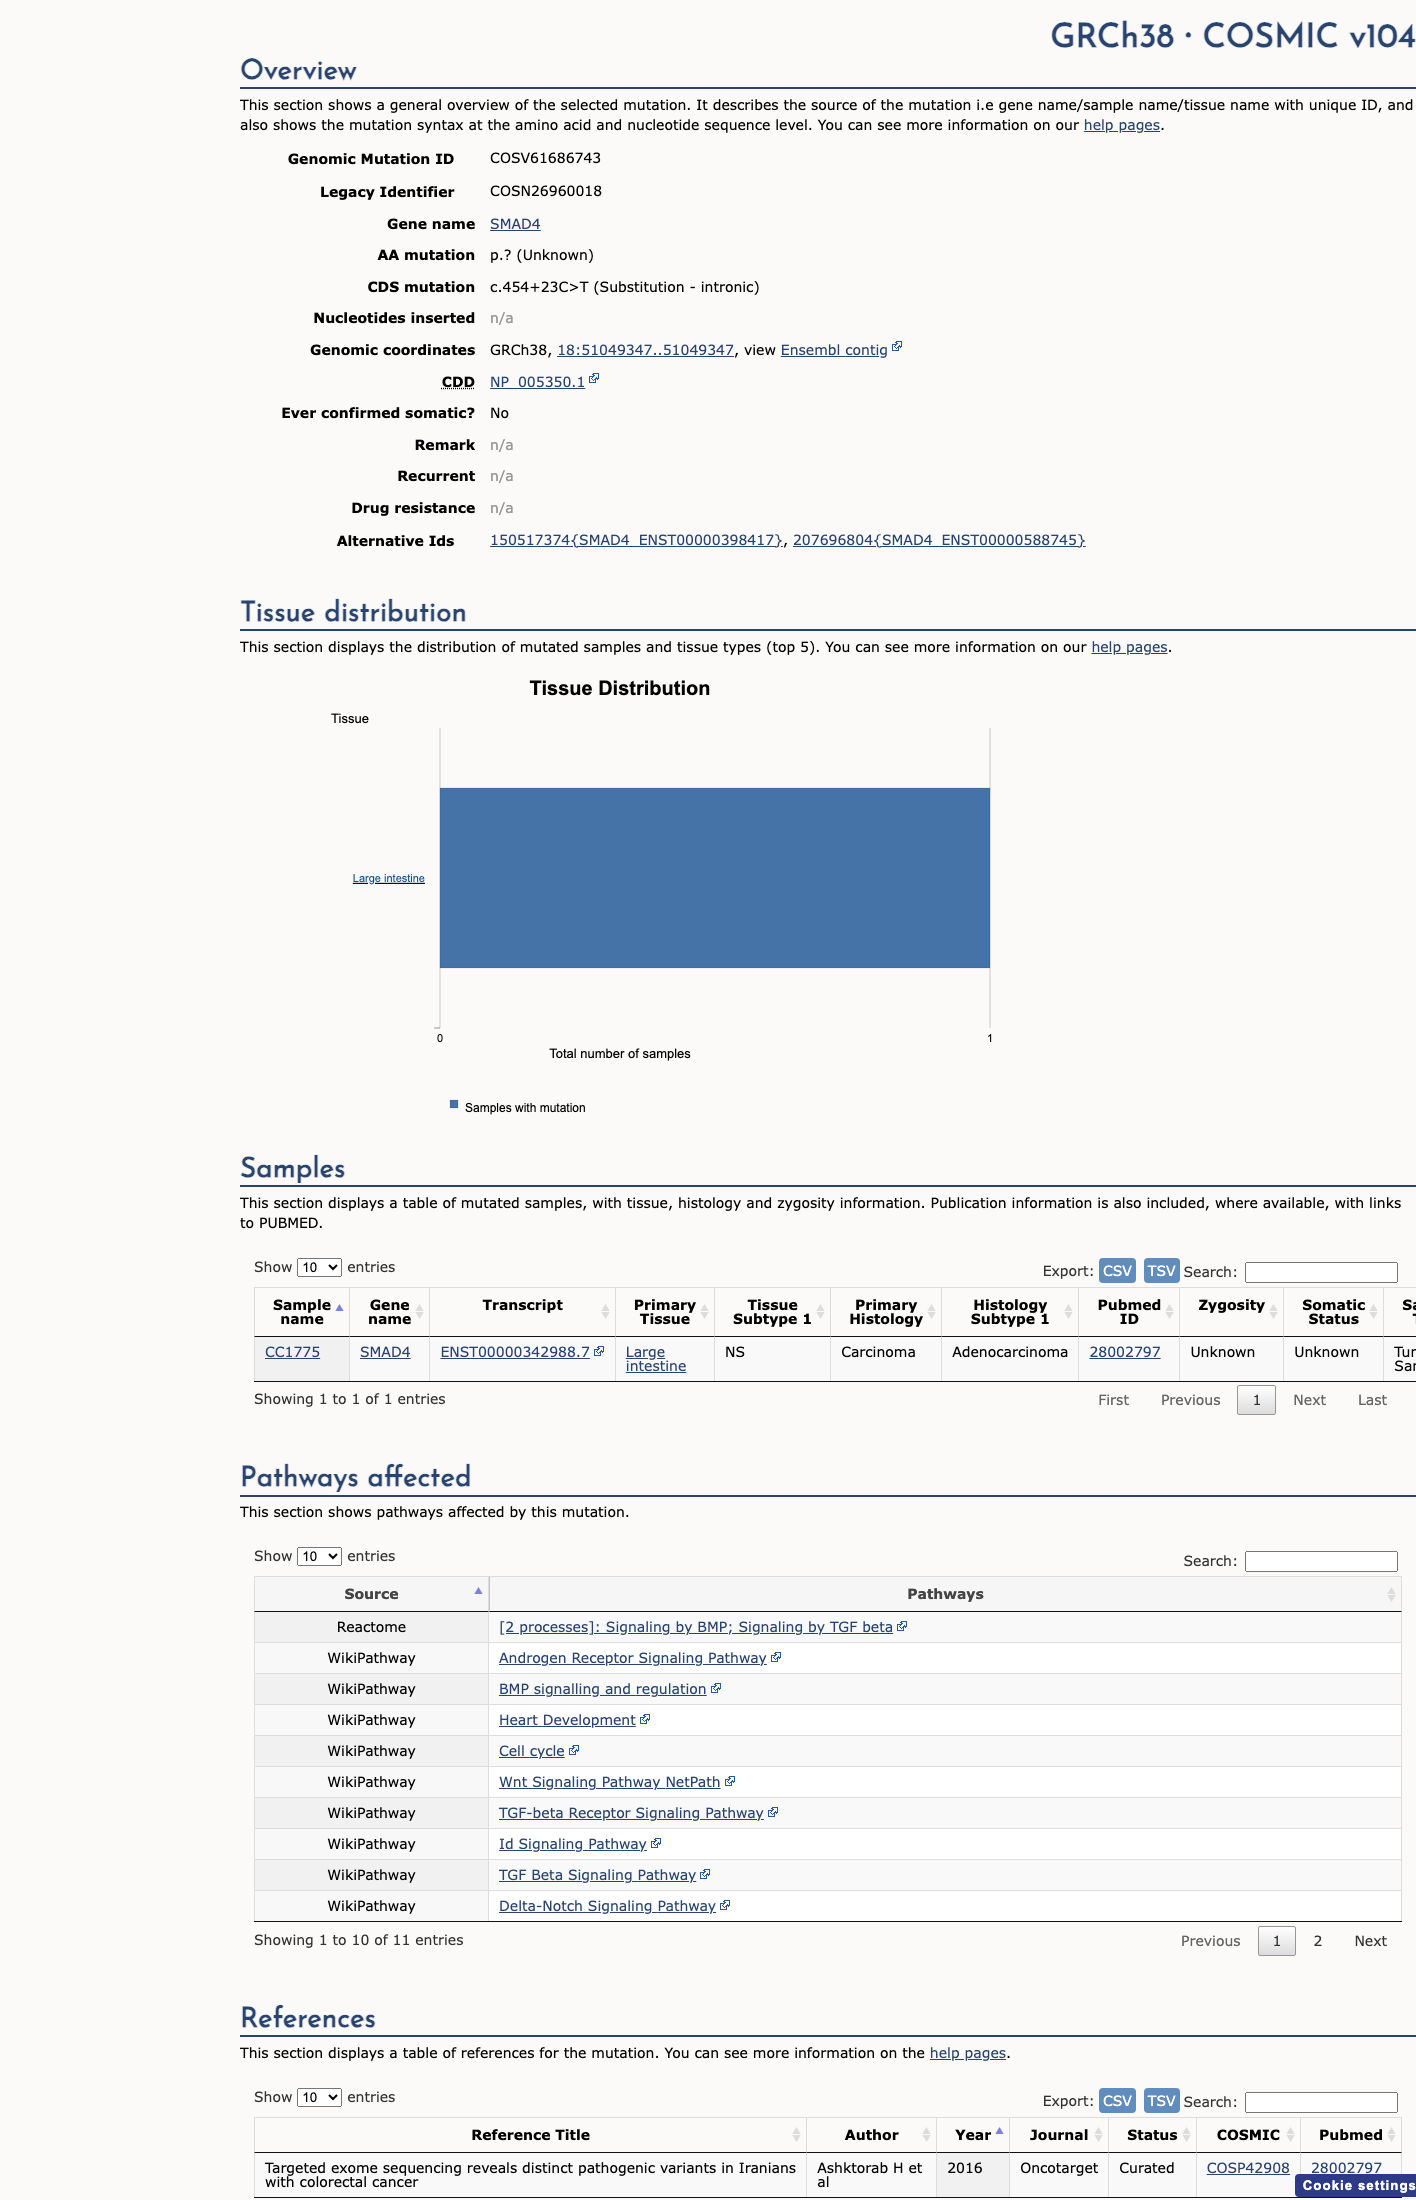
This variant has previously been reported in somatic cancers (COSMIC; COSV61686743, n = 1 times).
Hotspots
This variant does not lie in a statistically significant cancer hotspot.
COSMIC ↗
Sources & reference links
7Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Triaged references · 1 PMID not cited in assessment
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology. CLINVAR