León-Castillo et al. 2020 custom POLE framework · vleon-castillo-2020-custom-framework-v1
0%
complete
Re-run interpretation
Re-run this variant?
This variant was interpreted by pipeline
7.1.5.
The current version is
8.0.0.
Re-running queues a fresh interpretation; the result replaces this page when complete.
Re-runs are not free. Blank = draws on today's public interpretation allowance.
With a code = uses 1 of that code's uses.
POLE encodes the catalytic subunit of DNA polymerase epsilon, the enzyme that replicates the leading strand of DNA during cell division and participates in DNA repair. It contains a proofreading domain that corrects replication errors, keeping the accumulation of mutations in check. Germline mutations in POLE cause polyposis and predispose to colorectal cancer, and are also linked to a rare syndrome of facial dysmorphism, immunodeficiency, livedo, and short stature. Somatic mutations, particularly in the proofreading domain, occur in colorectal and endometrial cancers, where they drive an ultra-mutated tumor phenotype and are associated with better responses to immune checkpoint inhibitors.
This variant
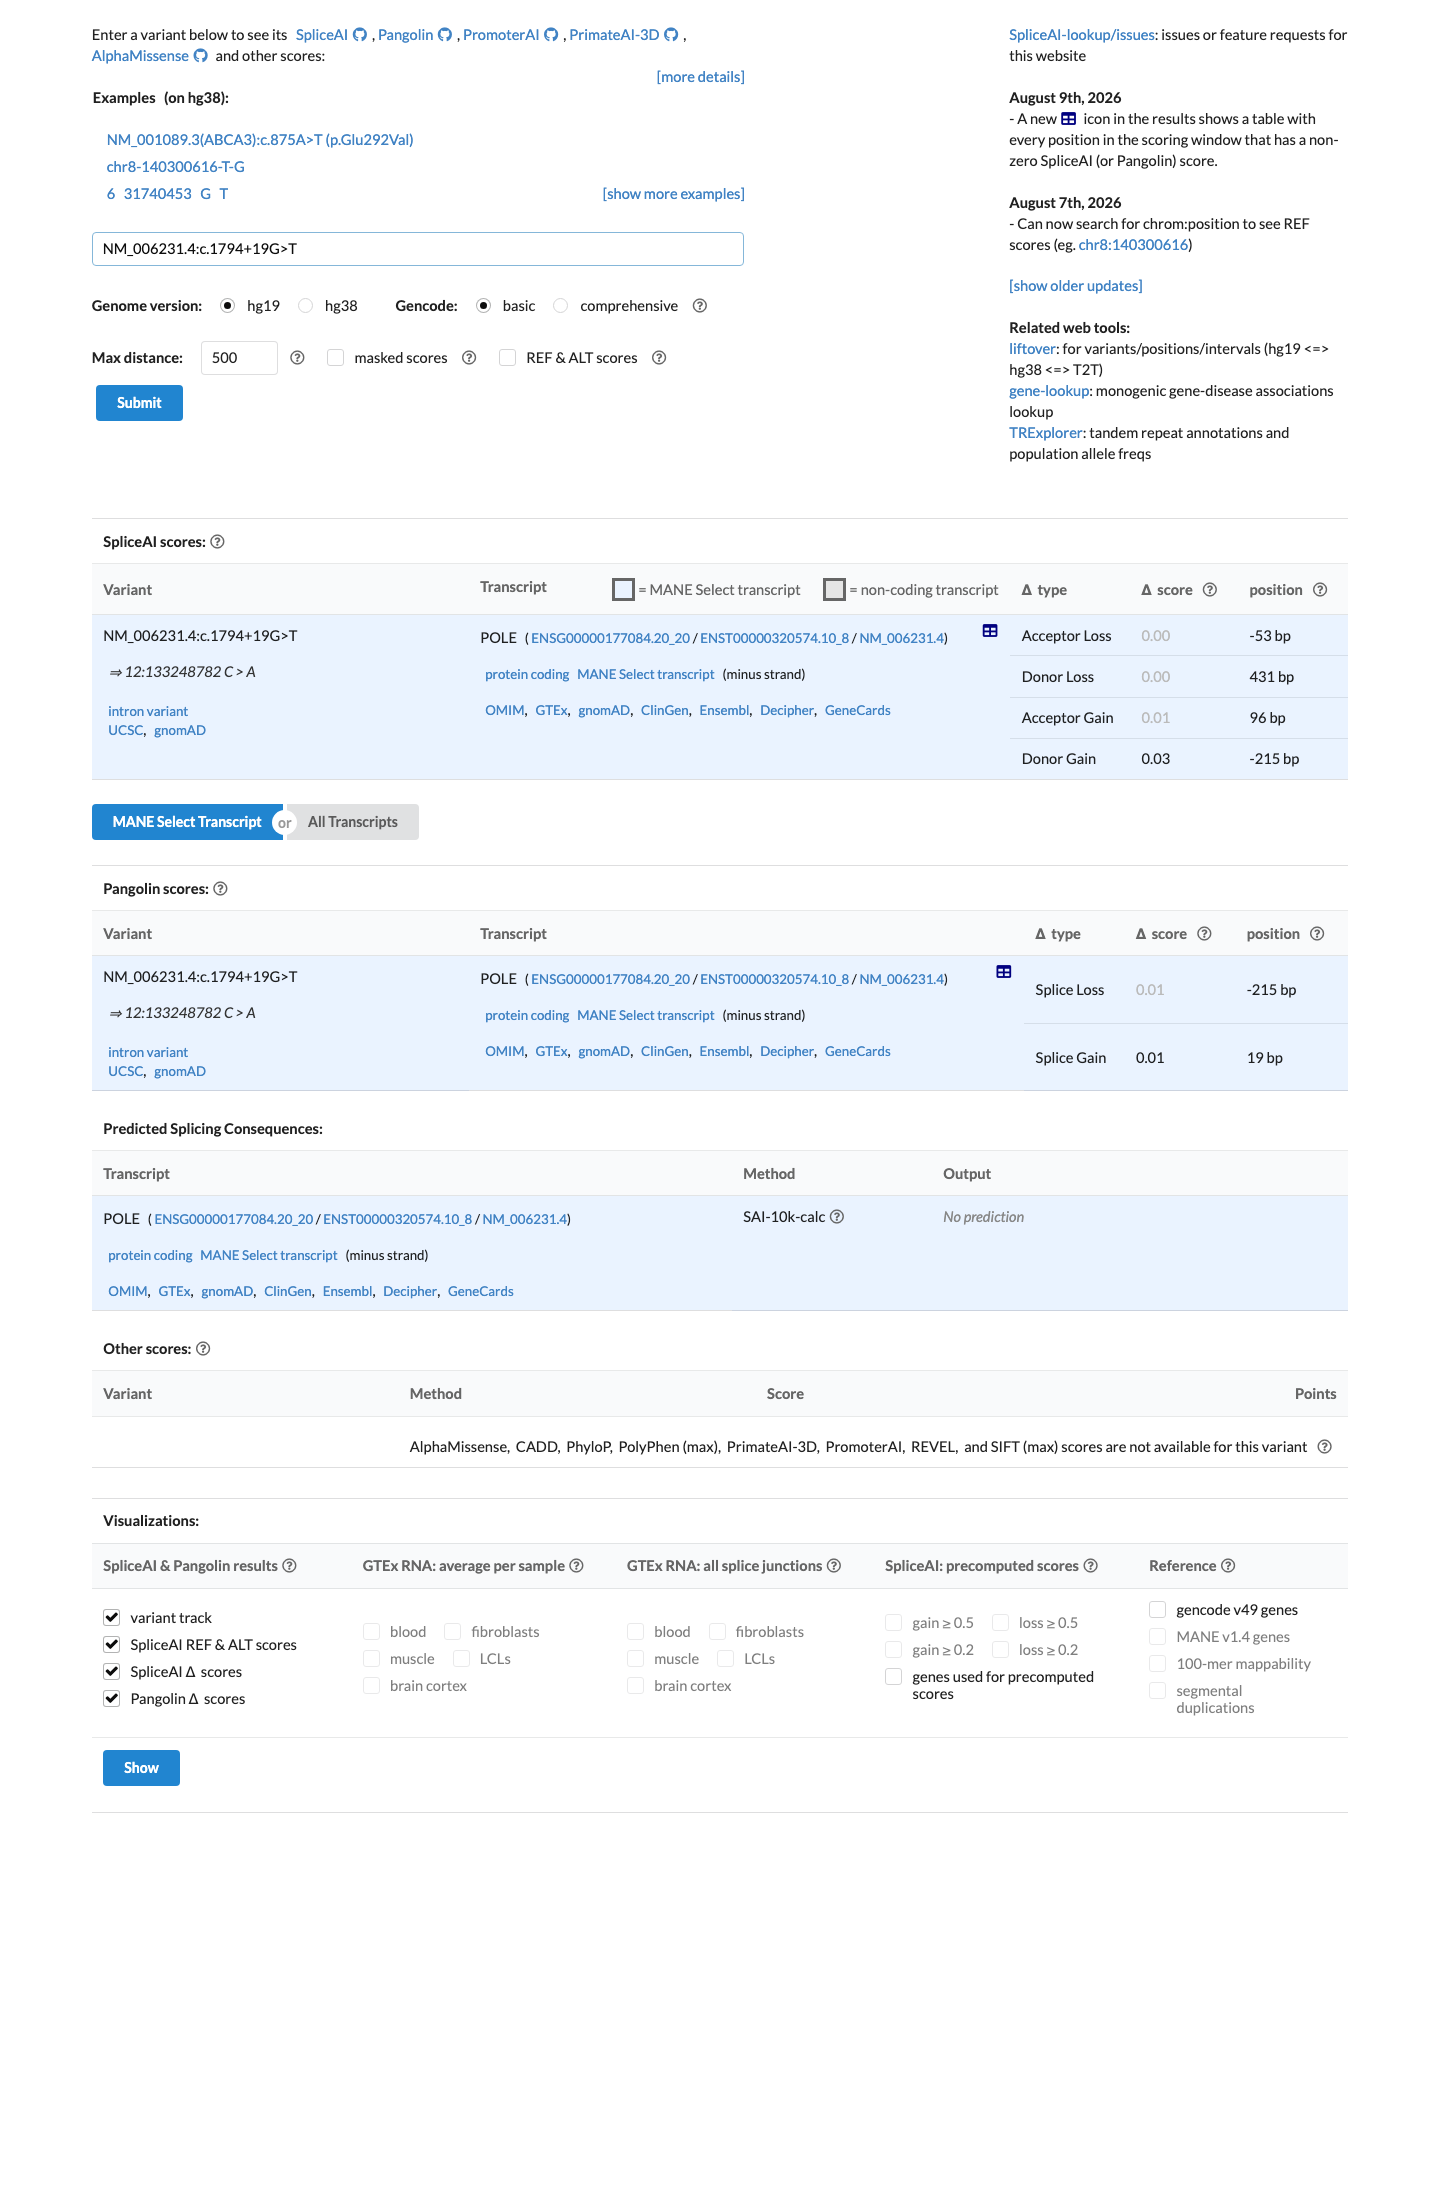
POLE's main disease mechanism is missense change in its proofreading domain, so a deep intronic substitution falls outside the recognized pathogenic pattern. With no predicted splice effect (SpliceAI max delta 0.031) yet essentially no population frequency (2/1,568,238 alleles), current evidence cannot determine whether this variant alters POLE function or colorectal cancer predisposition, hence VUS.
Transcript
NM_006231.4
HGVS · transcript:coding
NM_006231.4:c.1794+19G>T
GRCh38
chr12:132672196 C>A
GRCh37
chr12:133248782 C>A
Only PM2 (Supporting) and BP4 (Supporting) are met, which does not satisfy any pathogenic or benign ACMG/AMP combination, so the classification is VUS.
Classification rationale
PM2BP4VUS
POLE c.1794+19G>Tunknown · exon 16i
PM2 (Supporting): essentially absent from population databases - only 2/1,568,238 alleles in gnomAD v4.1 and none in gnomAD v2.1 or gnomAD-Canada v1.0, below the 0.1% rare-variant threshold. BP4 (Supporting): SpliceAI max delta 0.031 is below the <0.1 threshold, predicting no significant splice impact. Synthesis: with only PM2_Supporting and BP4_Supporting, no pathogenic or benign ACMG/AMP 2015 combination is reached, yielding a classification of VUS.
PM2 + BP4→VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria
may be applied incorrectly, sources may be misread, and a confident-looking
classification can still be wrong. Double-check every criterion and
its underlying evidence before relying on any call.
Gene diagram
· NM_006231.4 · variants mapped to exon structure
POLENM_006231.4
Fetching transcript structure from UCSC…
Exons
—
Transcript span
—
Strand
—
Variants mapped
—
Source
UCSC ncbiRefSeqCurated
All variants in POLE—click a row to locate it on the plot · use the link column to open its page
Protein
Location
Classification
Link
Applied criteria · 2 applied · 18 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
✓
PM2supportingPathogenic
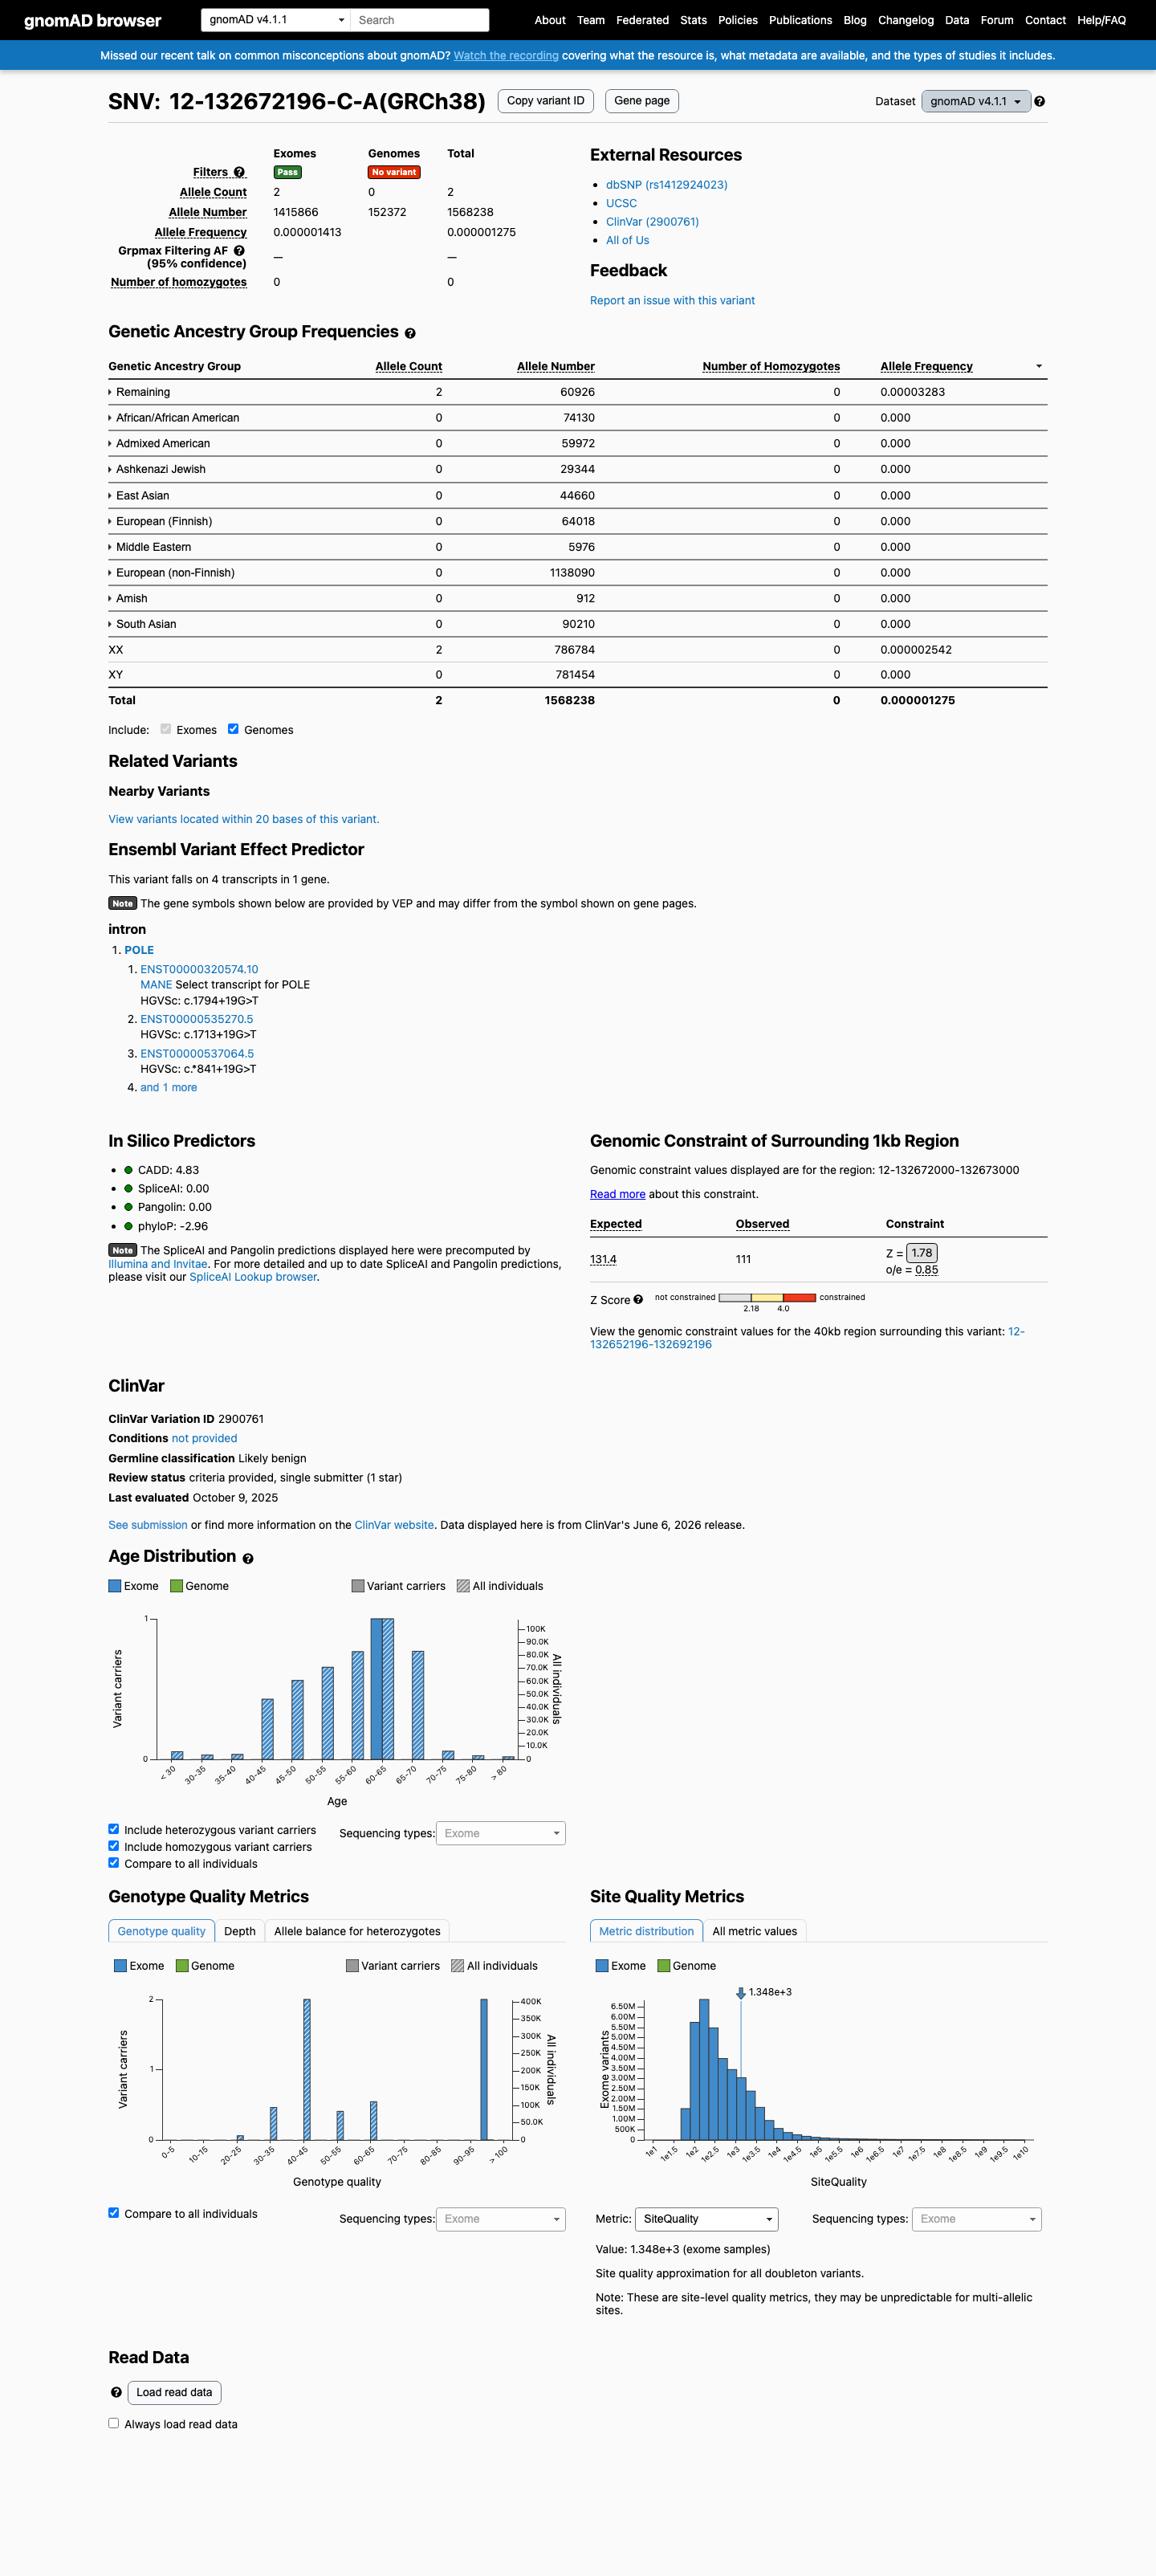
Met (Supporting): absent from gnomAD v2.1 and gnomAD-Canada v1.0, with only 2/1,568,238 alleles (AF 1.28e-06) in gnomAD v4.1, below the 0.1% rare-variant threshold.
The variant is absent from gnomAD v2.1.gnomAD v4.1 reports 2/1,568,238 total alleles, total AF 1.27532e-06, 0 homozygotes, and maximum reported subpopulation AF 3.28267e-05 from 2/60,926 alleles.The variant is absent from gnomAD-Canada v1.0.
Met (Supporting): SpliceAI max delta 0.031 is below the <0.1 BP4 threshold, predicting no significant splice impact.
The local POLE BP4 rule applies only to exact missense variants represented in Supplementary Table S2 or S3. The queried c.1794+19G>T intronic variant is absent from both reviewed tables, so the generic non-missense pathway applies.SpliceAI reports a maximum delta of 0.031 (DS_DG 0.031), below the generic BP4 threshold of 0.1 for intronic, synonymous, or non-canonical splice-position variants.
Assessed · not applied
· 6 not met · 12 not assessed
Pathogenic
PVS1Not met: deep intronic substitution with no defined protein consequence (p.?) and SpliceAI max delta 0.031, predicting no loss-of-function splice alteration.
PS2Not assessed: no parental testing results establish that the variant is absent from both biological parents (de novo).
PS3Not assessed: no validated functional assay evidence for this variant was available to demonstrate a damaging effect.
PS4Not assessed: the POLE framework's PS4 applies only to recurrent missense hotspots, and no case-control enrichment data exist for this intronic variant.
PM3Not assessed: no affected-proband observations, pathogenic allele in the same gene, or phase data establish a trans configuration.
PM6Not assessed: no parental testing or phenotype concordance data support a de novo occurrence.
PP1Not assessed: no pedigree or segregation data from affected or unaffected relatives were available.
PP3Not met: SpliceAI max delta 0.031 is below the >0.2 PP3 threshold, predicting no splice impact.
PP4Not assessed: no individual or tumor phenotype data establish a presentation highly specific to POLE-related disease.
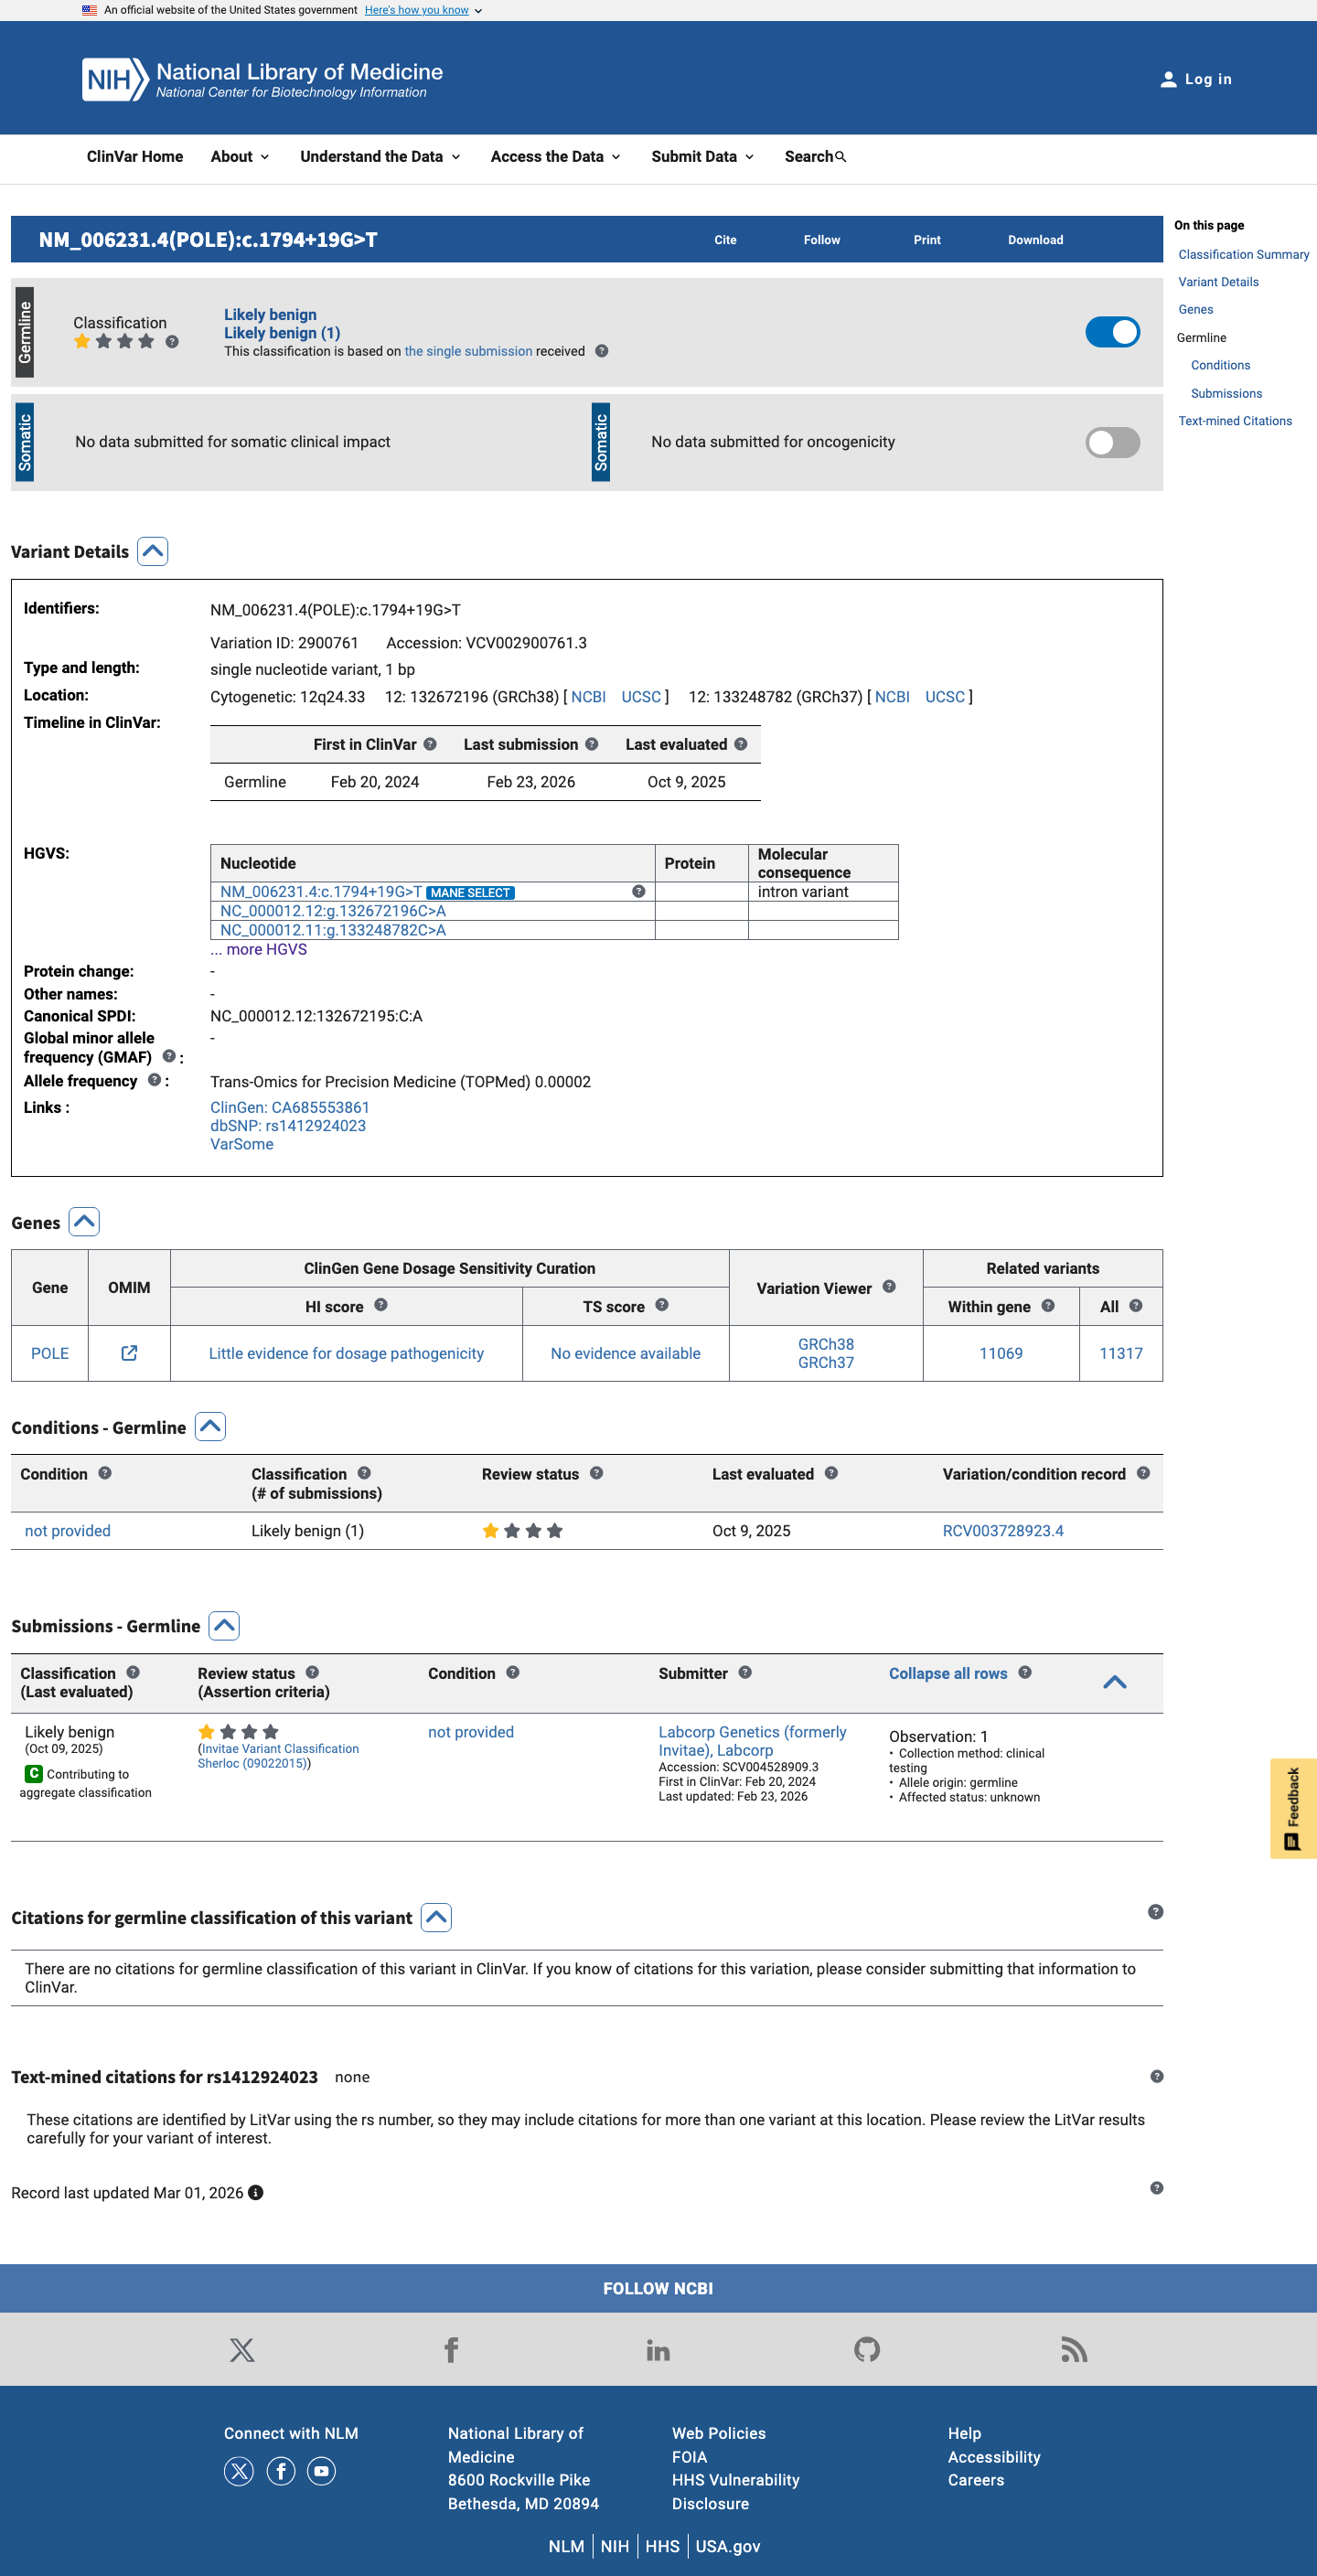
PP5Not met: the ClinVar record has no expert-panel submission, only a single clinical laboratory assertion.
Benign
BA1Not met: the highest allele frequency (3.28e-05) is far below the 1% BA1 threshold.
BS1Not met: the highest allele frequency (3.28e-05) is below the 0.3% BS1 threshold.
BS2Not assessed: no evidence documents carriers as phenotypically healthy adults, so benign homozygosity cannot be established.
BS3Not assessed: no functional assay evidence demonstrated a benign effect; the single-submitter Likely benign ClinVar entry is not functional evidence.
BS4Not assessed: no unaffected-carrier relatives or non-segregating meioses were documented.
BP2Not assessed: no co-occurrence of the variant with a pathogenic allele or cis/trans phase data were available.
BP5Not assessed: no carrier genotype or phenotype data show the phenotype is explained by an alternative molecular diagnosis.
BP6Not met: the single Likely benign laboratory submission cannot trigger BP6 because the record has no expert-panel submission.
This variant is present in gnomAD v4.1 (AF= 1.27532e-06; MAF= 0.00013%, 2/1568238 alleles, homozygotes = 0) and has highest observed frequency in the Remaining individuals population (AF= 3.28267e-05; MAF= 0.00328%, 2/60926 alleles, homozygotes = 0).
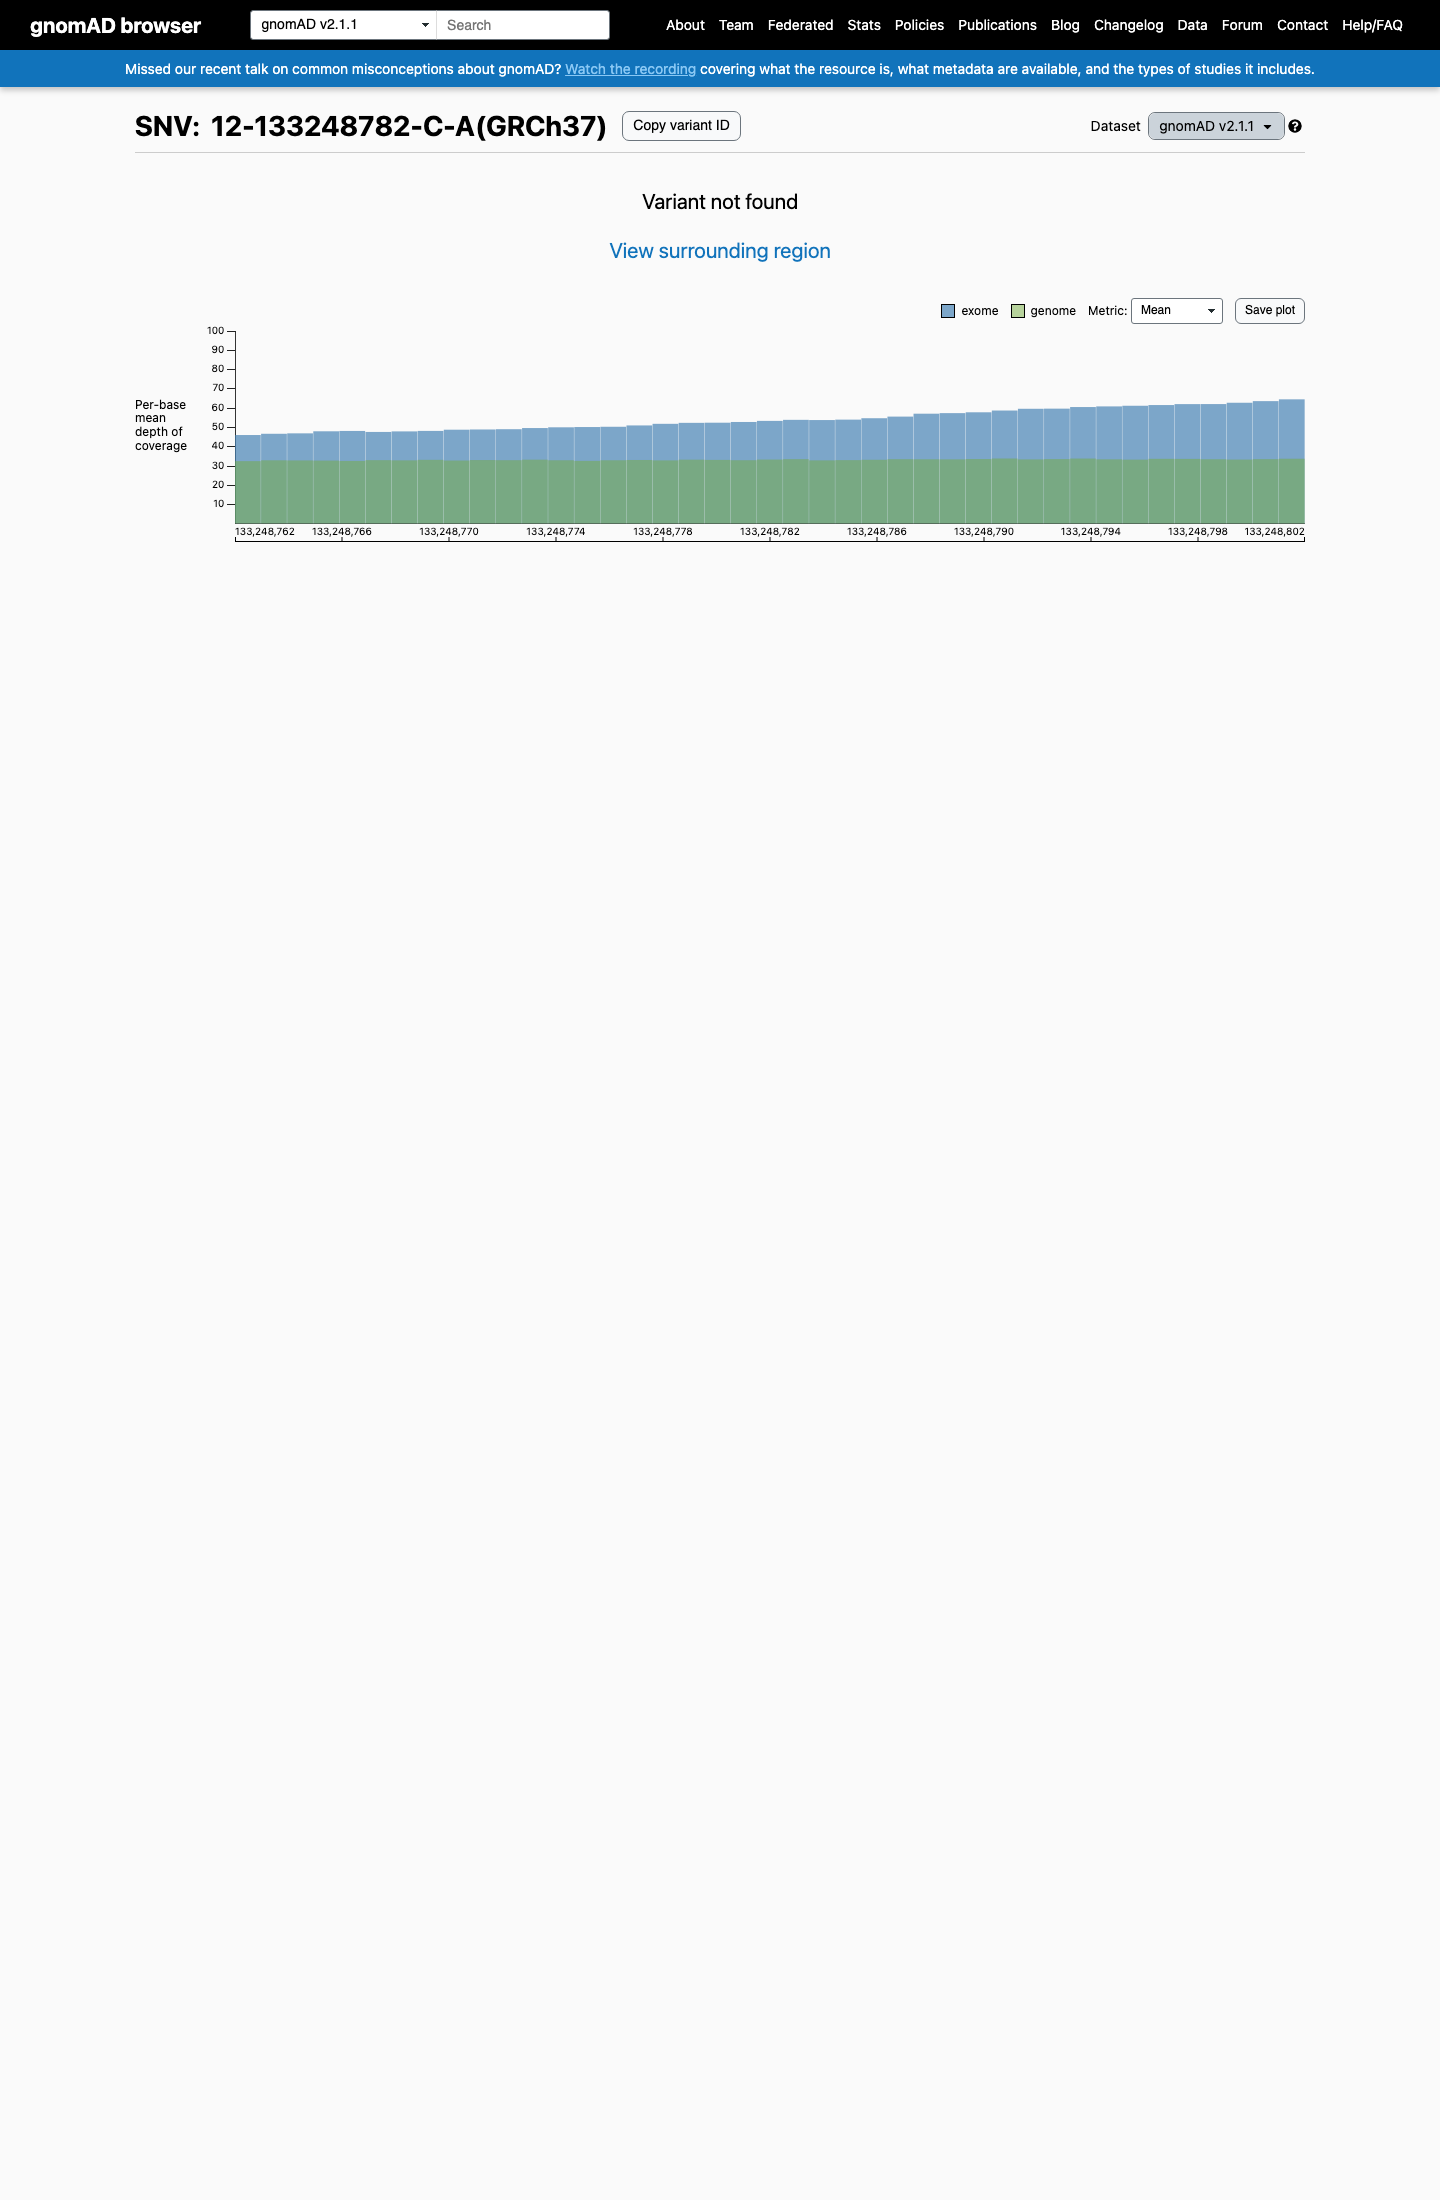
v2.1
Absent from gnomAD v2.1.
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.00013%
· 2 / 1,568,238
0 hom
Remaining individuals
2 / 60,926
0.0033%
+ 9 not observed (Admixed American, European (Finnish), Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American, European (non-Finnish))