Back
NM_006231.4:c.1924-5C>T
p.? · POLE
León-Castillo et al. 2020 custom POLE framework · vleon-castillo-2020-custom-framework-v1
0%
complete
Final classification
VUS
PM2BP4
POLE
c.1924-5C>T
p.?
unknown · exon 17i

POLE encodes the catalytic subunit of DNA polymerase epsilon, the enzyme that replicates the leading strand of DNA during cell division and participates in DNA repair. It contains a proofreading domain that corrects replication errors, keeping the accumulation of mutations in check. Germline mutations in POLE cause polyposis and predispose to colorectal cancer, and are also linked to a rare syndrome of facial dysmorphism, immunodeficiency, livedo, and short stature. Somatic mutations, particularly in the proofreading domain, occur in colorectal and endometrial cancers, where they drive an ultra-mutated tumor phenotype and are associated with better responses to immune checkpoint inhibitors.

This variant

The variant lies in POLE, which encodes the DNA polymerase epsilon catalytic subunit involved in leading-strand replication and DNA repair and is implicated in inherited polyposis and colorectal-cancer predisposition.

Transcript
NM_006231.4
HGVS · transcript:coding
NM_006231.4:c.1924-5C>T
GRCh38
chr12:132668742 G>A
GRCh37
chr12:133245328 G>A
VUS: PM2 supporting evidence for rarity conflicts with BP4 supporting computational benign evidence, and no POLE framework or ACMG/AMP combination rule is satisfied.
Classification rationale
PM2 BP4 VUS
POLE c.1924-5C>T unknown · exon 17i

PM2 supporting: the variant is extremely rare in gnomAD, with non-cancer AF 6.75776e-06. BP4 supporting: SpliceAI maximum delta 0.084 is below the <=0.1 benign threshold.

PM2 + BP4 → VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_006231.4 · variants mapped to exon structure
POLE NM_006231.4
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 18 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
PM2 supporting Pathogenic
Met at supporting strength: gnomAD v3.1 non-cancer AF is 6.75776e-06, below the <=0.0001 PM2 threshold.
gnomAD v2.1 non-cancer reports the variant as absent.gnomAD v3.1 non-cancer reports 1/147978 genome alleles, AF 6.757761288840233e-06, and zero homozygotes.Default all-comers gnomAD v2.1 reports AF 7.959692119108834e-06 and gnomAD v4.1 reports AF 4.337470040474792e-06; both report zero homozygotes.
BP4 supporting Benign
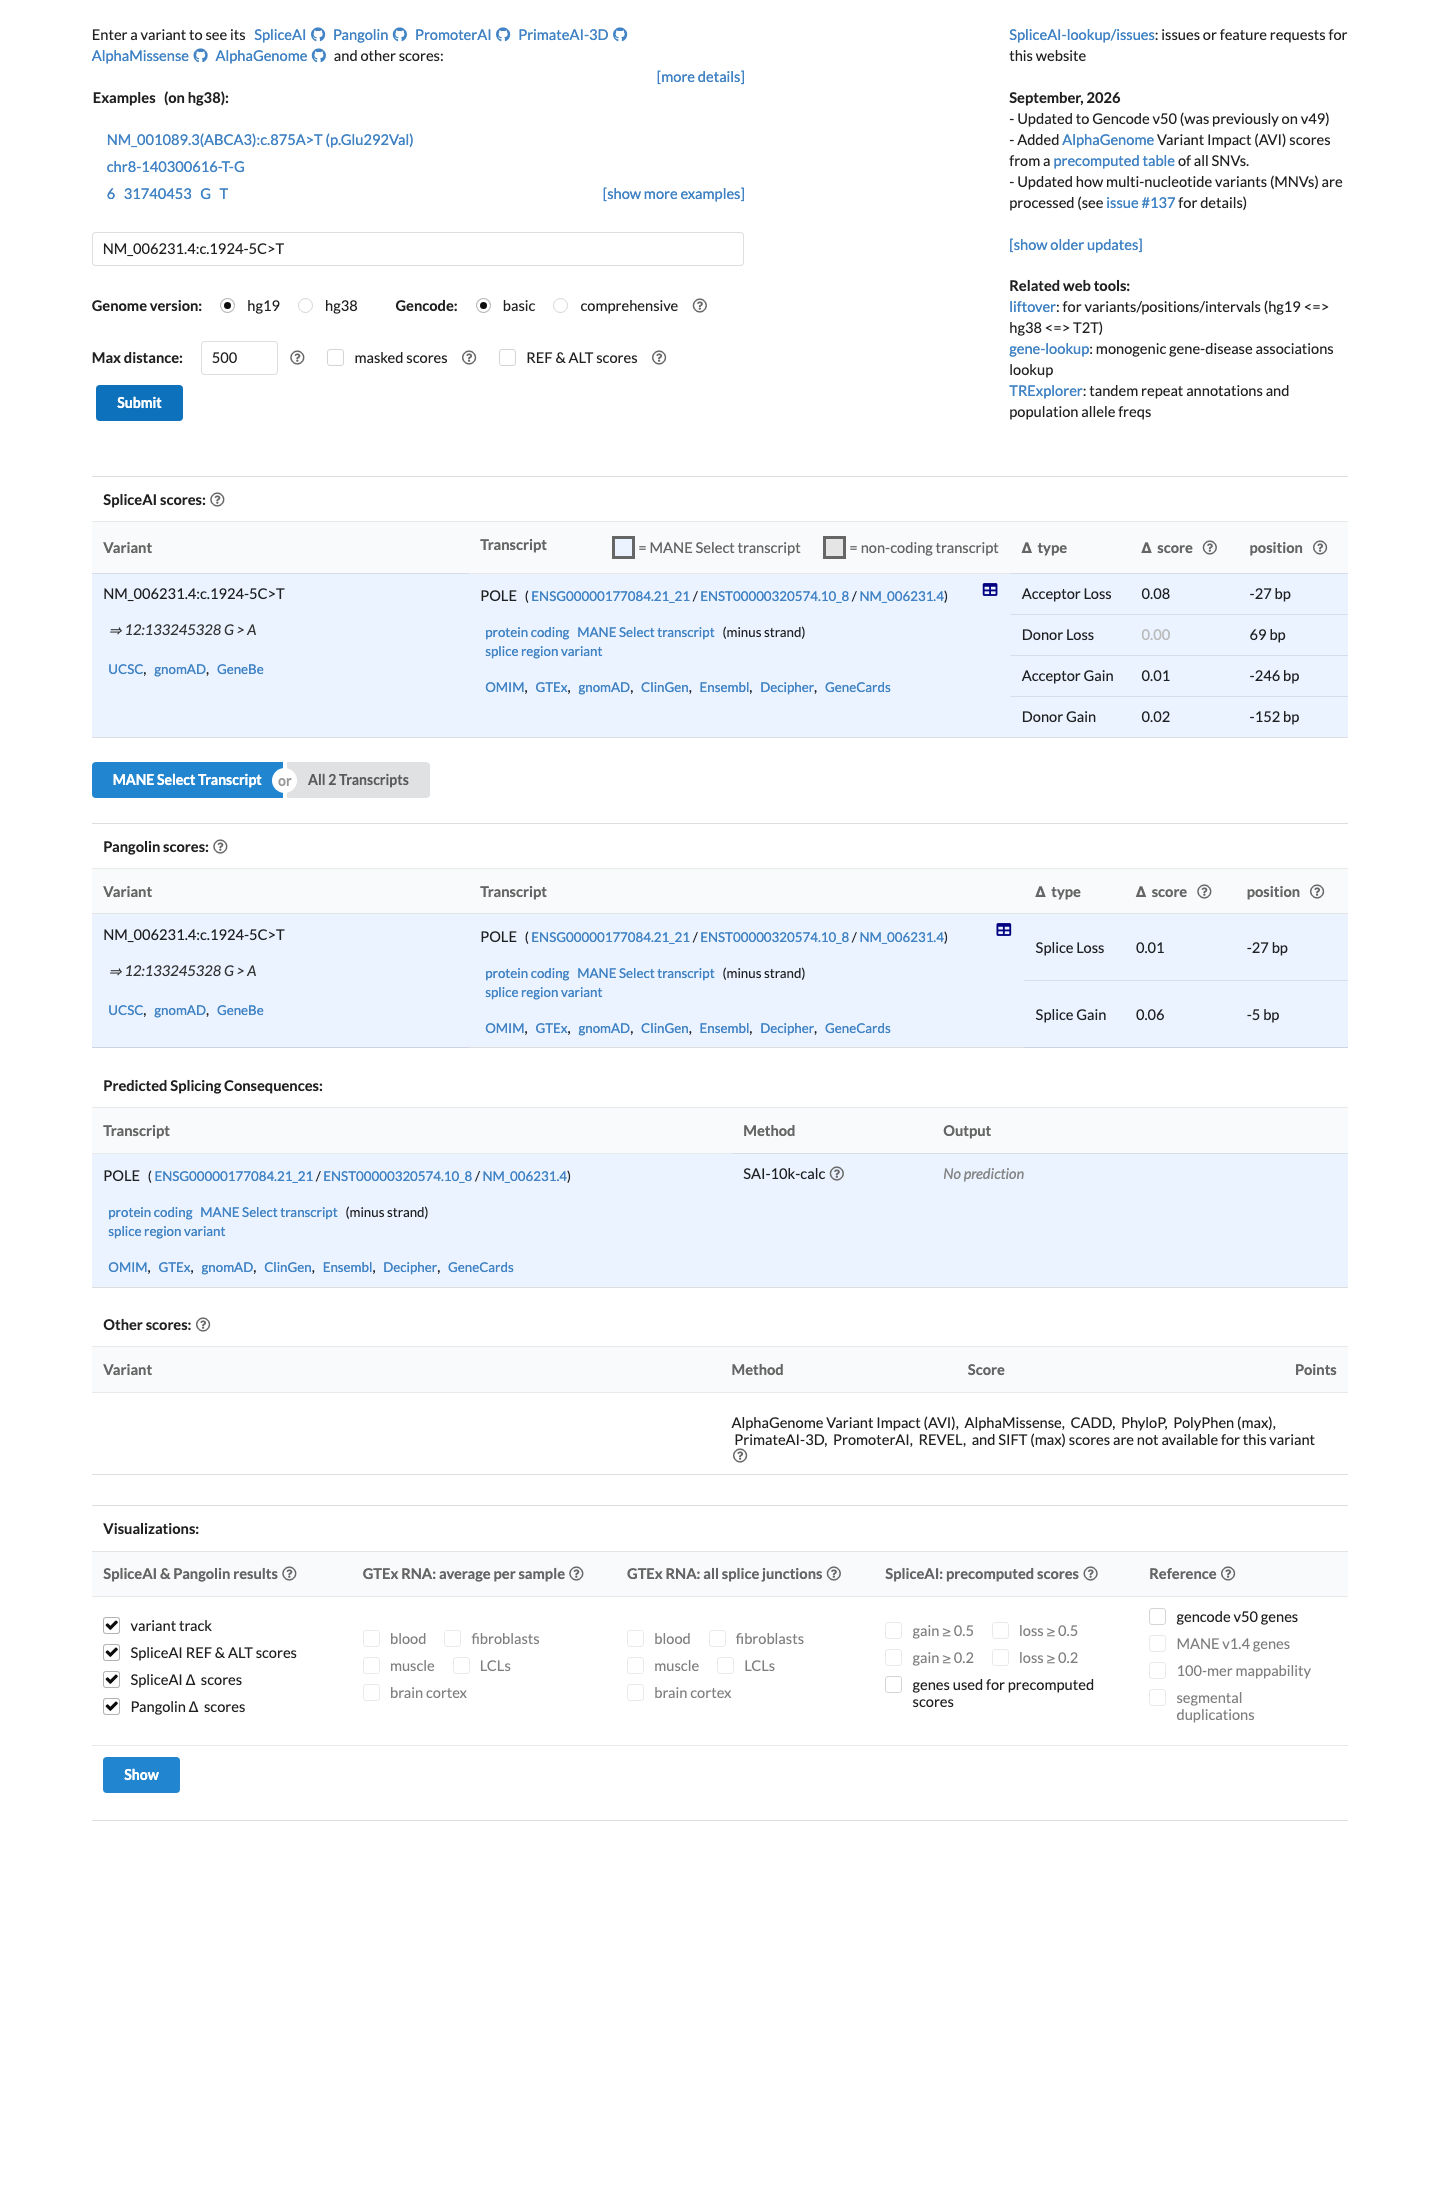
Met, supporting: SpliceAI maximum delta 0.084 meets the <=0.1 BP4 supporting threshold.
The variant is intronic/splice-region, so SpliceAI is the sole applicable PP3/BP4 predictor path; REVEL is not applicable.SpliceAI reported DS_AG 0.007, DS_AL 0.084, DS_DG 0.017, and DS_DL 0.000; maximum delta 0.084.The generic SpliceAI BP4 supporting cutoff is <=0.1, from Jaganathan et al. 2019 (PMID:30661751); 0.084 meets it.
Assessed · not applied · 7 not met · 11 not assessed
Pathogenic
PVS1 Not met: c.1924-5C>T is a splice-region SNV at -5, outside the canonical ±1,2 PVS1 null-variant positions, with no demonstrated abnormal transcript or NMD.
PS2 Not assessed: no proband-parent genotypes, confirmed parentage, or documented absence of the variant in both parents are available.
PS3 Not assessed: no controlled RNA, minigene, protein, or other validated functional assay is reported for c.1924-5C>T; SpliceAI 0.084 is computational, not PS3 evidence.
PS4 Not met: the exact splice-region variant is absent from the governing recurrence tables and is not one of the specified pathogenic POLE missense hotspots.
PM3 Not assessed: no affected-proband observation, pathogenic second allele, or phase evidence establishes this variant in trans for PM3.
PM6 Not assessed: no clinical report describes this variant as de novo with unconfirmed parentage or provides supporting proband and parental data.
PP1 Not assessed: no affected relatives, informative meioses, or family segregation results are reported for this variant.
PP3 Not met: SpliceAI maximum delta 0.084 is below the >=0.2 PP3 supporting threshold.
PP4 Not assessed: no patient-level, highly specific POLE-related phenotype is available for comparison with the PP4 requirement.
PP5 Not met: ClinVar has zero expert-panel submissions and no exact-variant expert-panel Pathogenic or Likely pathogenic classification.
Benign
BA1 Not met: the highest reported allele frequency is 0.000201694, far below the 0.05 BA1 threshold.
BS1 Not met: the highest reported allele frequency is 0.000201694, below the generic 0.01 BS1 threshold.
BS2 Not assessed: gnomAD shows zero homozygotes, but no documented unaffected carrier or healthy-adult observation is available for BS2.
BS3 Not assessed: no controlled assay demonstrates normal splicing or protein function for c.1924-5C>T, and SpliceAI 0.084 cannot establish BS3.
BS4 Not assessed: no affected, well-phenotyped individual lacking the variant or documented non-segregation analysis is available.
BP2 Not assessed: no documented co-occurring pathogenic variant or cis/trans phase result supports BP2 for this variant.
BP5 Not assessed: no governing BP5 specification or validated BP5-specific evidence is available for this variant.
BP6 Not met: ordinary Likely benign laboratory submissions cannot trigger BP6 because ClinVar reports zero expert-panel submissions.
N/A · 8 PS1 · PM1 · PM4 · PM5 · PP2 · BP1 · BP3 · BP7
Research & evidence
Population frequency
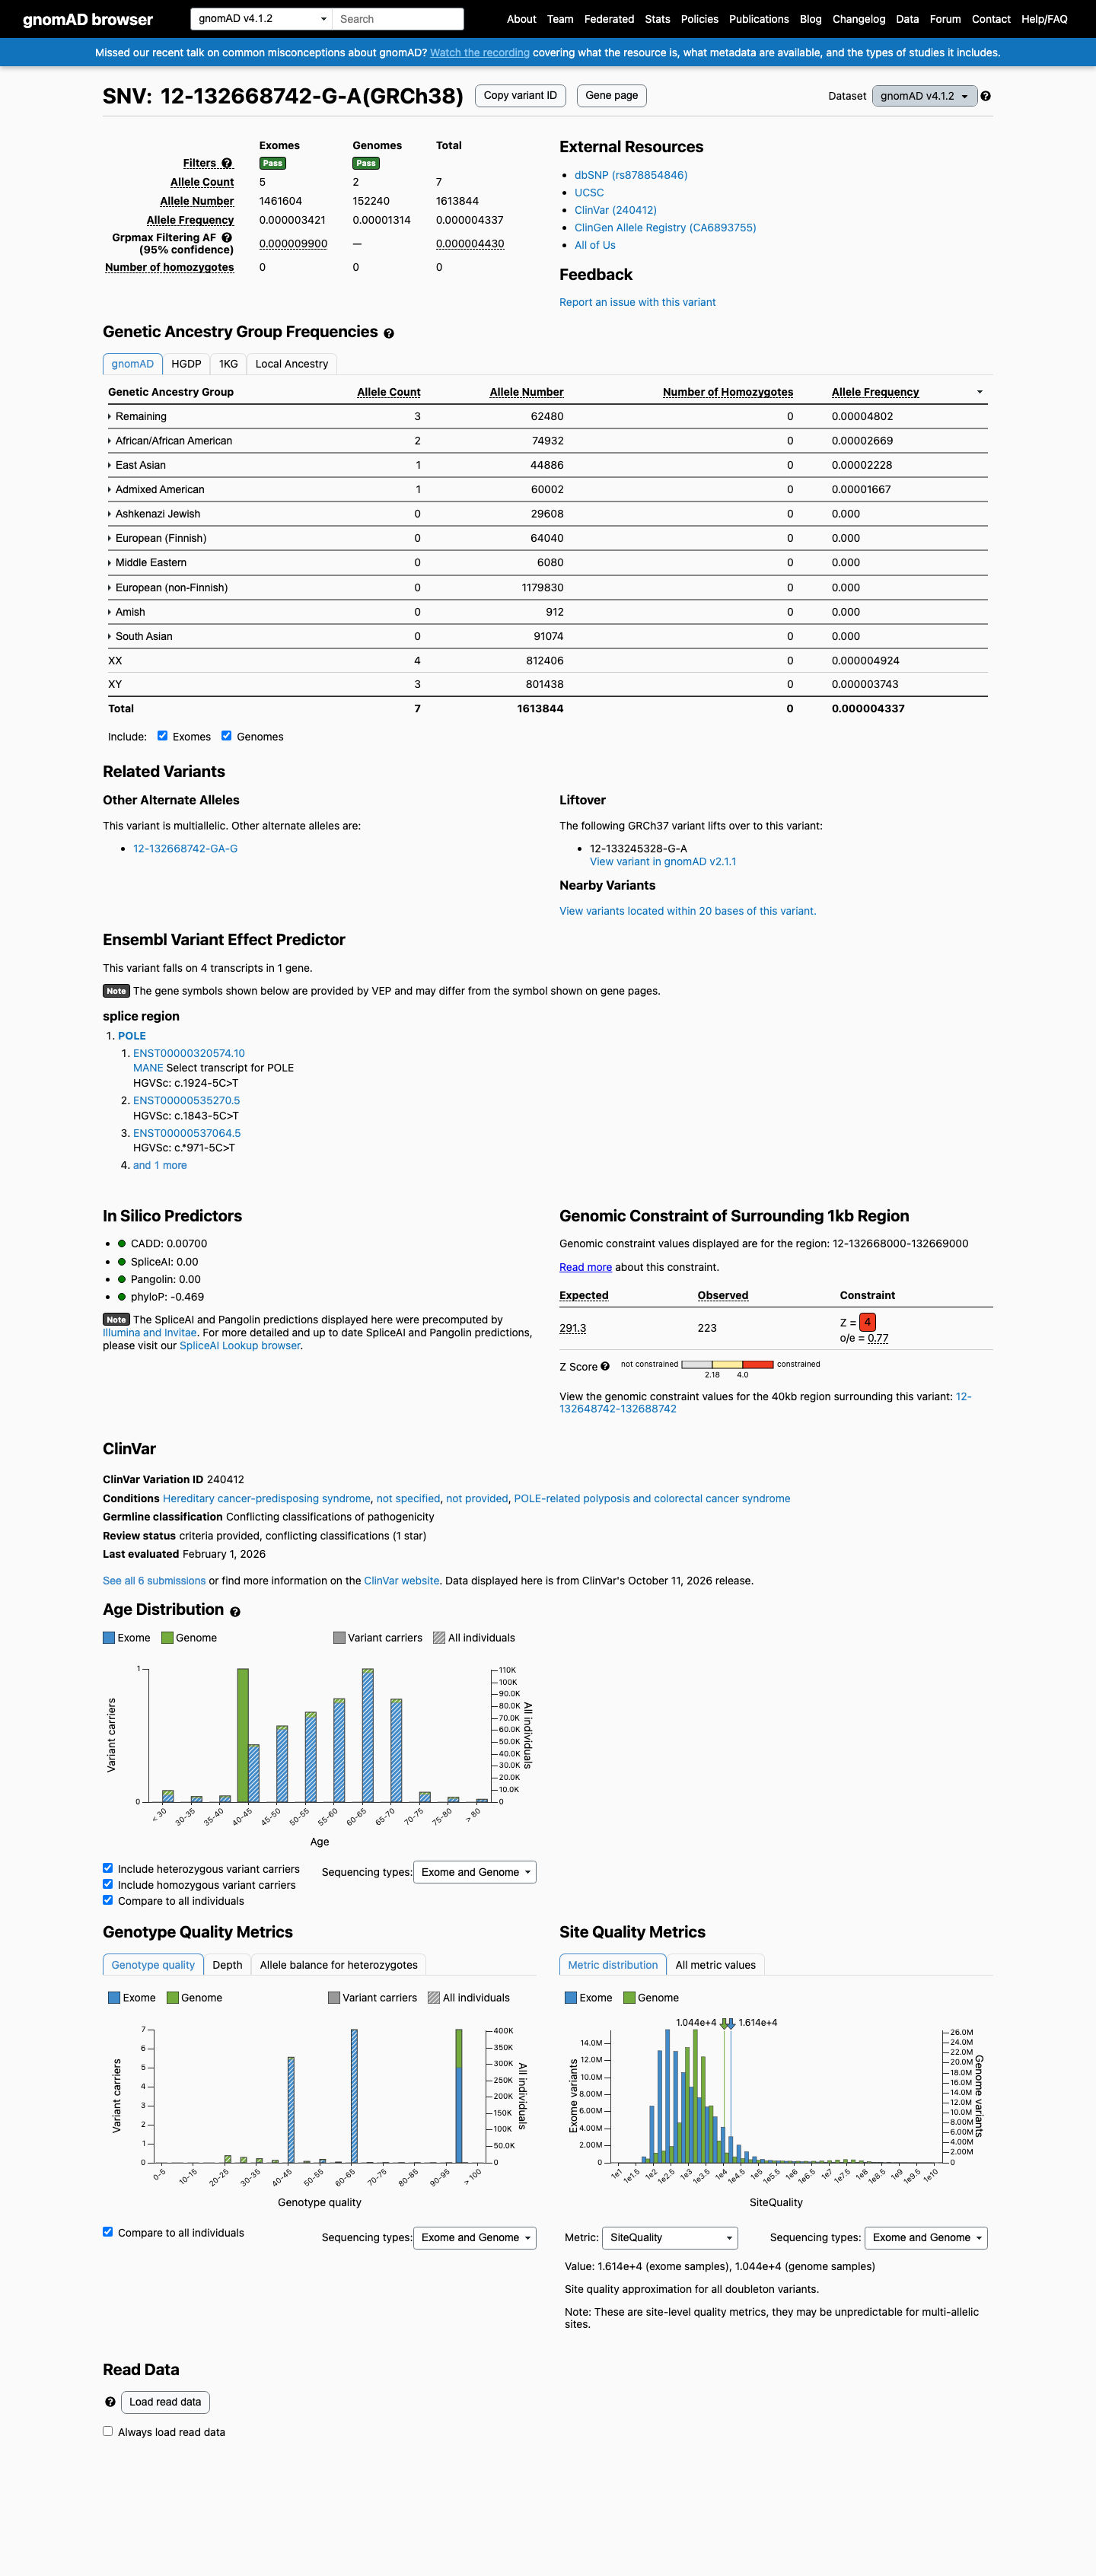
gnomAD v4.1 screenshot
gnomAD v4.1
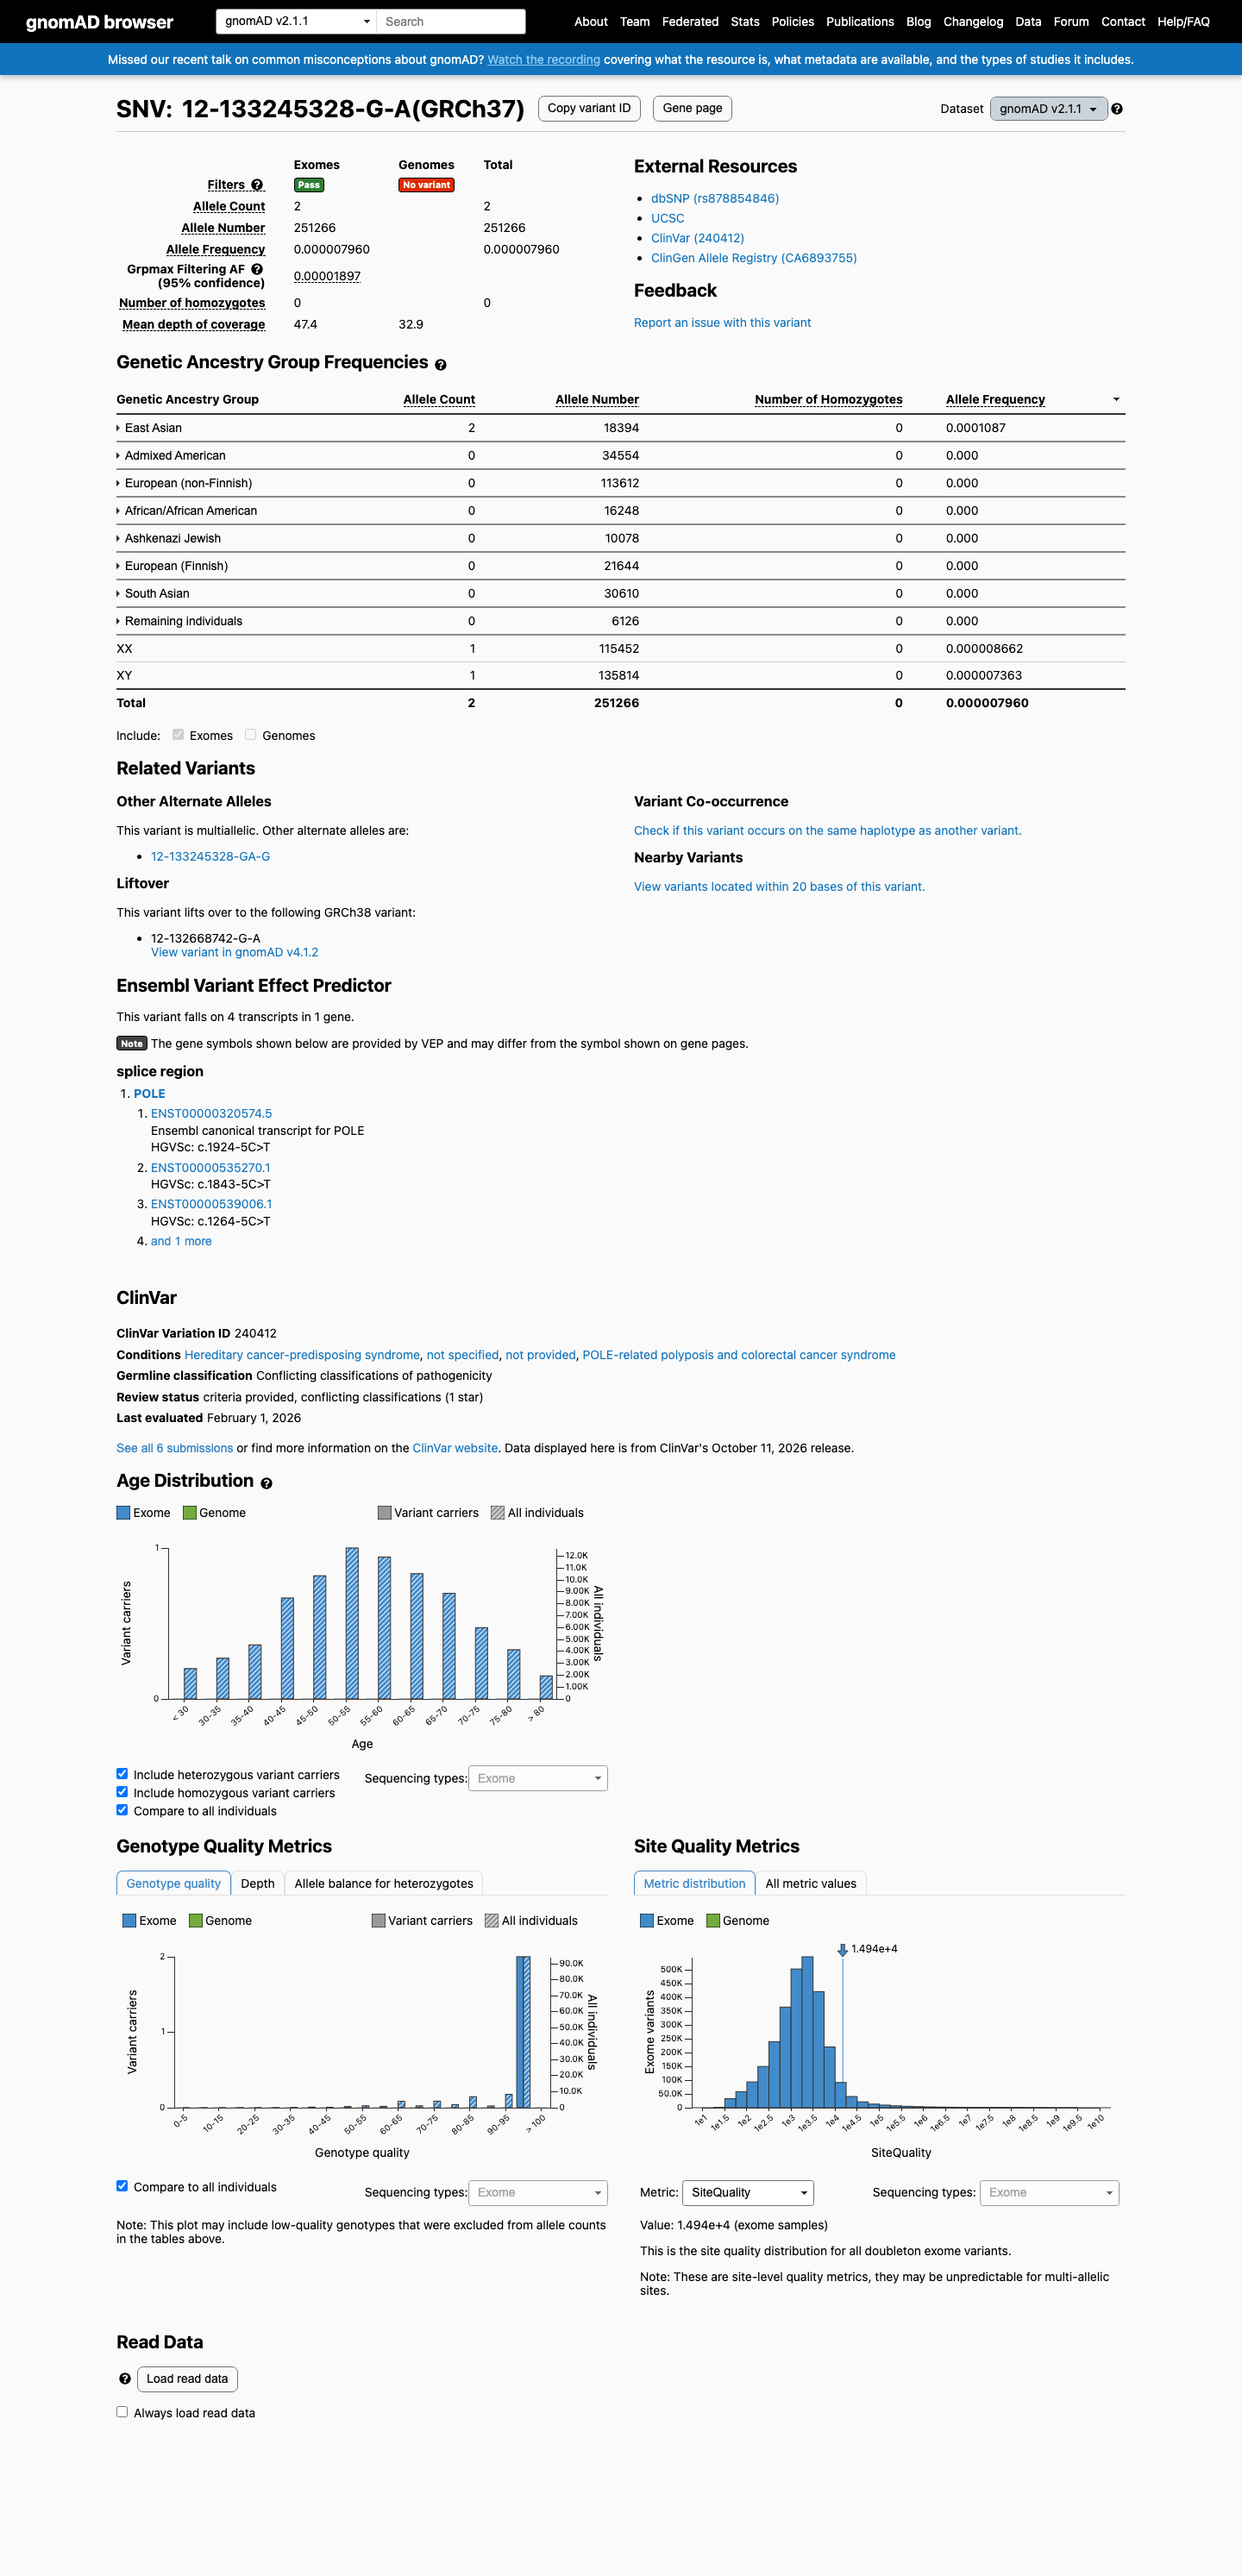
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 4.33747e-06; MAF= 0.00043%, 7/1613844 alleles, homozygotes = 0) and has highest observed frequency in the Remaining individuals population (AF= 4.80154e-05; MAF= 0.00480%, 3/62480 alleles, homozygotes = 0); grpmax FAF= 4.43e-06.
v2.1
This variant is present in gnomAD v2.1 (AF= 7.95969e-06; MAF= 0.00080%, 2/251266 alleles, homozygotes = 0) and has highest observed frequency in the East Asian population (AF= 0.000108731; MAF= 0.01087%, 2/18394 alleles, homozygotes = 0); grpmax FAF= 1.897e-05.
🇨🇦 CA
This variant is present in gnomAD-Canada v1.0 (AF= 0.00032583903551645487, 6/18414 alleles, homozygotes = 0).
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.00043% · 7 / 1,613,844
0 hom · FAF 0.00044%
Remaining individuals
3 / 62,480
0.0048%
African/African American
2 / 74,932
0.0027%
East Asian
1 / 44,886
0.0022%
Admixed American
1 / 60,002
0.0017%
+ 6 not observed (European (Finnish), Amish, Middle Eastern, South Asian, Ashkenazi Jewish, European (non-Finnish))
gnomAD v2.1
0.0008% · 2 / 251,266
0 hom · FAF 0.0019%
East Asian
2 / 18,394
0.011%
+ 7 not observed (African/African American, Admixed American, Ashkenazi Jewish, European (Finnish), European (non-Finnish), Remaining individuals, South Asian)
gnomAD Canada 🇨🇦
0.033% · 6 / 18,414
0 hom · FAF 0.2%
East Asian
6 / 1,338
0.45%
+ 8 not observed (African/African American, Latino/Admixed American, Ashkenazi Jewish, European (Finnish), Middle Eastern, European (non-Finnish), Remaining individuals, South Asian)
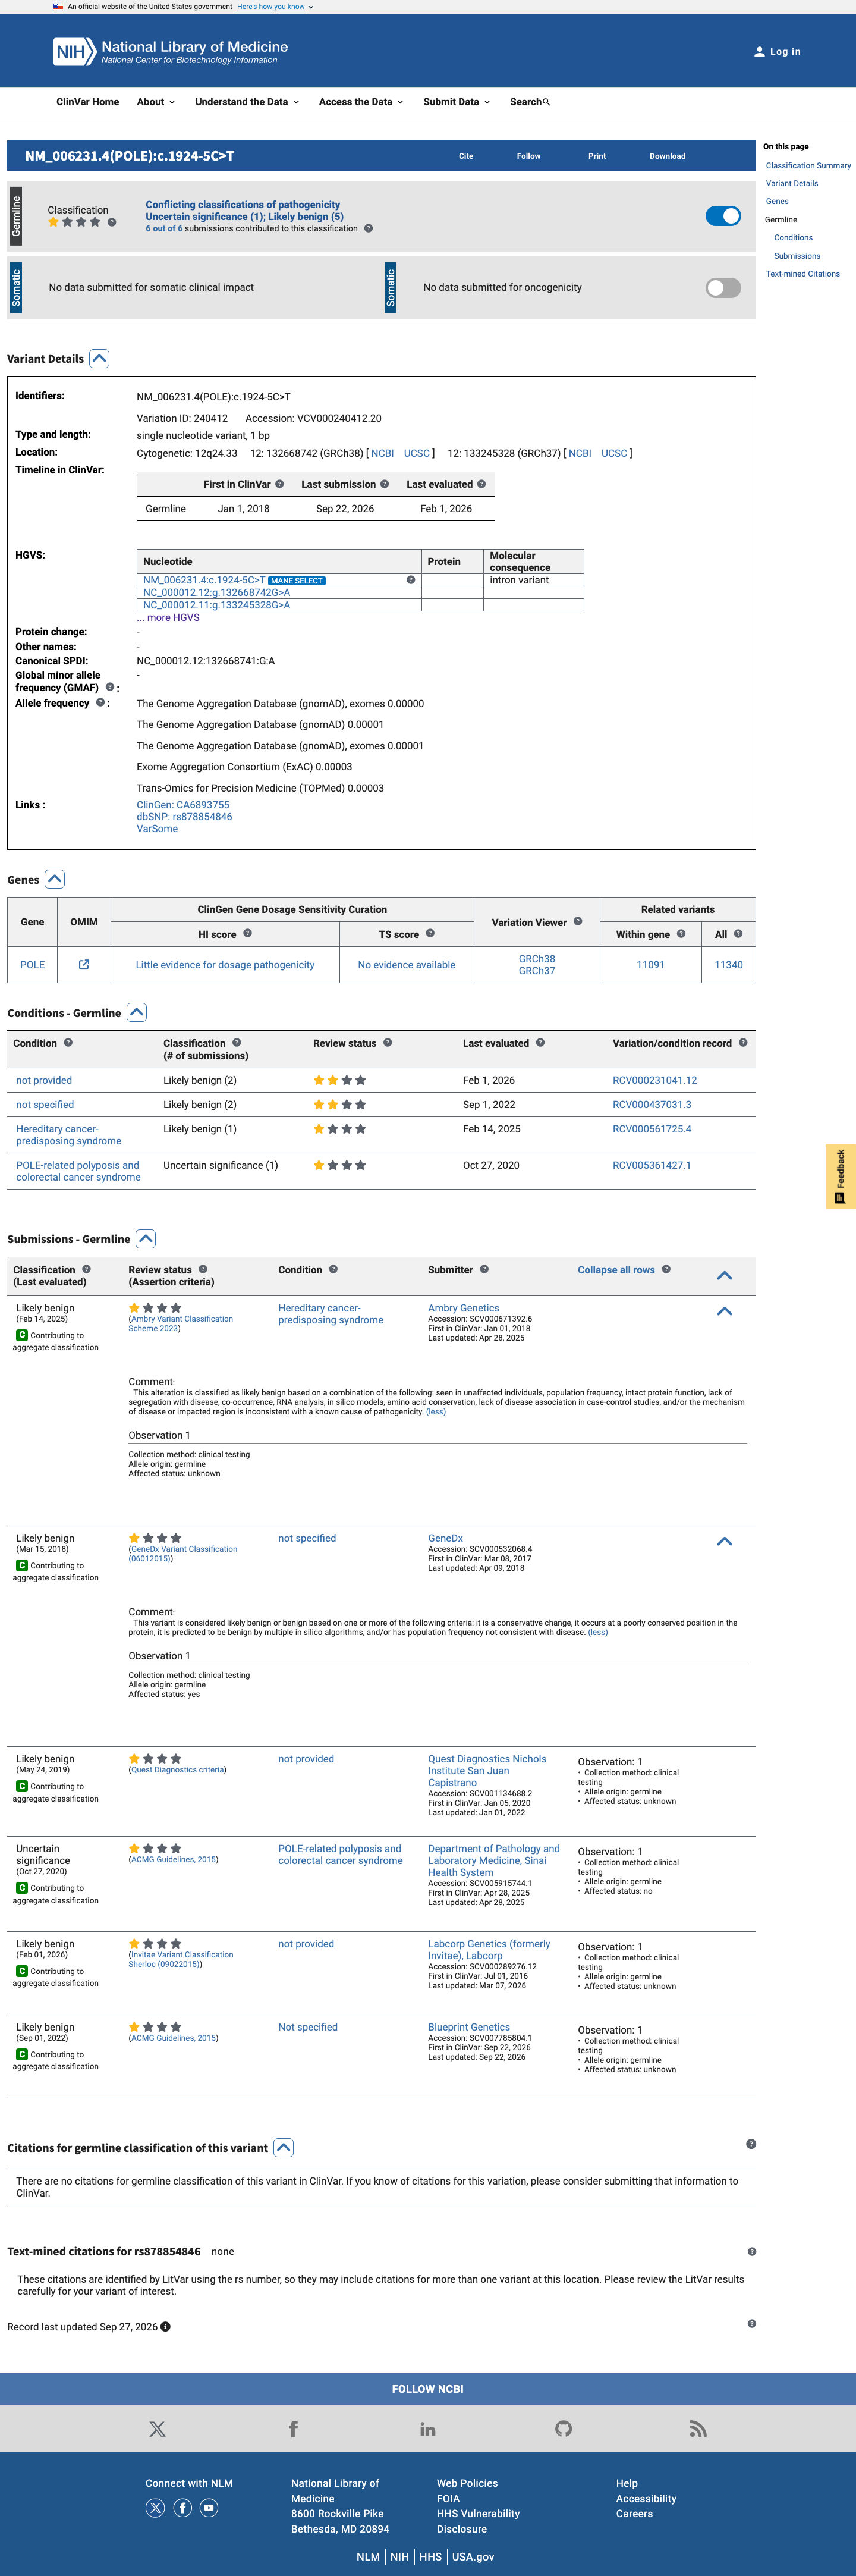
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Likely benign (5 clinical laboratories) and as Uncertain significance (1 clinical laboratory). (ClinVarID = 240412)
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.08).
Functional No data
No calibrated functional assay or RNA evidence was identified for this variant.
OncoKB ↗
COSMIC screenshot
COSMIC
Somatic evidence
COSMIC
This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant cancer hotspot.
COSMIC ↗
Literature · how each cited paper was used
1papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 3 further PMIDs triaged but not cited — see Sources & references.
Rule & framework references · cited for criterion definitions, not variant evidence
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.
Sources & reference links
7Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Triaged references · 3 PMIDs not cited in assessment
26467025 ↗ A Standardized DNA Variant Scoring System for Pathogenicity Assessments in Mendelian Disorders. CLINVAR
25394175 ↗ A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment. CLINVAR
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification criteria. CLINVAR