Back
NM_006231.4:c.2173+23G>T
p.? · POLE
León-Castillo et al. 2020 custom POLE framework · vleon-castillo-2020-custom-framework-v1
0%
complete
Final classification
VUS
BP4
POLE
c.2173+23G>T
p.?
unknown · exon 19i

POLE encodes the catalytic subunit of DNA polymerase epsilon, the enzyme that replicates the leading strand of DNA during cell division and participates in DNA repair. It contains a proofreading domain that corrects replication errors, keeping the accumulation of mutations in check. Germline mutations in POLE cause polyposis and predispose to colorectal cancer, and are also linked to a rare syndrome of facial dysmorphism, immunodeficiency, livedo, and short stature. Somatic mutations, particularly in the proofreading domain, occur in colorectal and endometrial cancers, where they drive an ultra-mutated tumor phenotype and are associated with better responses to immune checkpoint inhibitors.

This variant

This deep-intronic substitution lies well outside the POLE exonuclease proofreading domain whose defects drive polymerase proofreading-associated polyposis and colorectal cancer predisposition, and it produces no predicted change to splicing or to the polymerase epsilon protein, so its clinical significance in that gene context remains unresolved.

Transcript
NM_006231.4
HGVS · transcript:coding
NM_006231.4:c.2173+23G>T
GRCh38
chr12:132668333 C>A
GRCh37
chr12:133244919 C>A
VUS: only BP4 at supporting strength is met (SpliceAI max delta 0.003), one criterion short of the two supporting-benign criteria needed for Likely Benign.
Classification rationale
BP4 VUS
POLE c.2173+23G>T unknown · exon 19i

BP4 supporting: the deep-intronic change has SpliceAI max delta 0.003, below the 0.1 no-impact cutoff, so no splice effect is predicted.

BP4 → VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_006231.4 · variants mapped to exon structure
POLE NM_006231.4
Fetching transcript structure from UCSC…
Applied criteria · 1 applied · 20 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 1
Strength Supporting Moderate Strong Very strong
BP4 supporting Benign
Met at supporting: SpliceAI max delta 0.003 for this intronic variant, below the <=0.1 BP4 threshold.
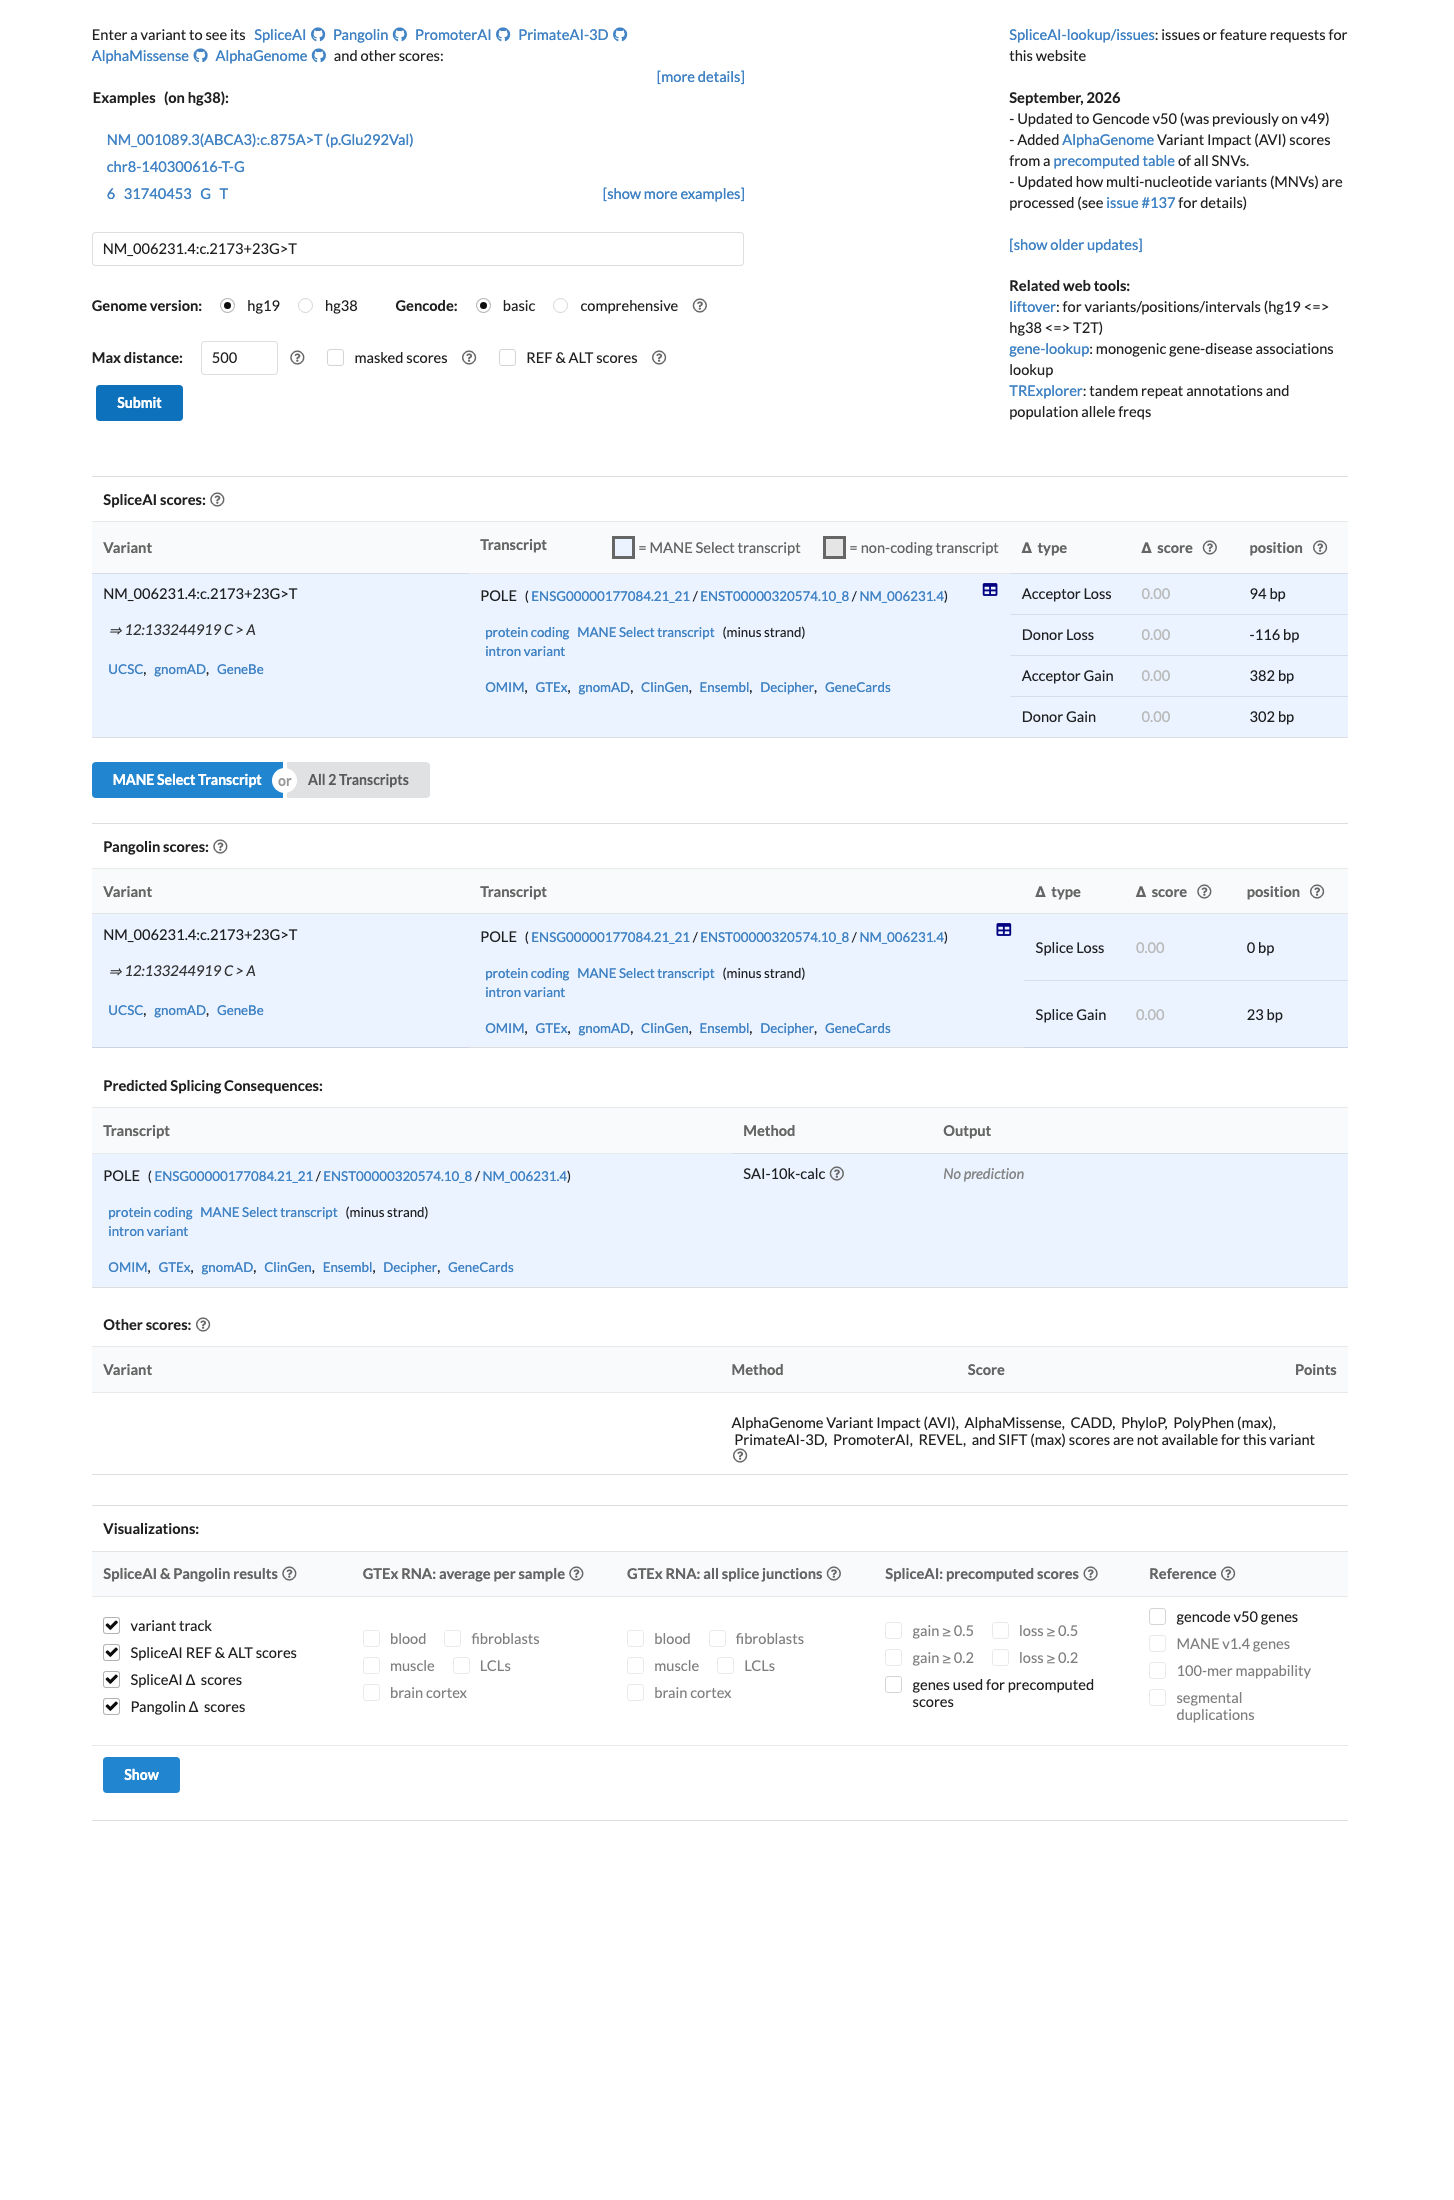
SpliceAI Lookup (hg37) for NM_006231.4:c.2173+23G>T returned a genuine, non-missing result (search_status result_found, spliceai_available=true, variant_id 12-133244919-C-A): DS_AG 0.00, DS_AL 0.003, DS_DG 0.00, DS_DL 0.001, max delta score 0.003, with predicted delta positions DP_AG 382, DP_AL 94, DP_DG 302, DP_DL -116.SpliceAI BP4 cutoff used verbatim as supplied: <=0.1 (supporting) indicates no meaningful splice impact (Jaganathan et al. 2019, Cell; PMID:30661751). 0.003 is well below it.Pangolin (SG 0.001, SL -0.001) from the same lookup is concordant with no splice-altering effect; used as corroboration only.
Assessed · not applied · 10 not met · 10 not assessed
Pathogenic
PVS1 Not met: this deep-intronic substitution (23 bp into intron 19) has SpliceAI max delta 0.003, so no null consequence is predicted and PVS1 cannot be triggered.
PS2 Not assessed: no proband or parental genotype data exist in this case to establish de novo occurrence.
PS3 Not assessed: no functional or RNA assay of POLE c.2173+23G>T exists; the only splice datum is computational (SpliceAI max delta 0.003).
PS4 Not met: variant recurrence count 0 in the COSMIC/TCGA EC table vs the PS4_Supporting requirement of combined count >=10.
PM1 Not met: intronic c.2173+23G>T (intron 19, p.?) lies outside the POLE exonuclease domain and all framework hotspots, and is present in gnomAD at 0.066% overall.
PM2 Not met: total allele frequency 0.000665 (gnomAD v2.1, 155/233,160 alleles) exceeds the 0.0001 PM2 rarity threshold in every dataset assessed.
PM3 Not assessed: no phase, pedigree, or second-variant data exist to determine whether a pathogenic POLE allele is in trans.
PM6 Not assessed: no proband or parental testing data are available to support an assumed de novo occurrence.
PP1 Not assessed: no pedigree or relative genotype data are available to assess co-segregation.
PP3 Not met: SpliceAI max delta 0.003 for this intronic variant, far below the >=0.2 supporting PP3 threshold.
PP4 Not assessed: no proband phenotype, HPO terms, or family history anywhere in the case, so PP4's patient-specific specificity requirement cannot be evaluated.
PP5 Not met: no ClinVar expert-panel assertion exists (0 expert-panel submissions); the sole record is Likely benign, single submitter.
Benign
BA1 Not met: highest ancestry frequency 0.59% (gnomAD v2.1 African/African American, 144/24,282 alleles) versus the 0.05 BA1 stand-alone threshold.
BS1 Not met: highest ancestry frequency 0.98% (gnomAD-Canada African/African American) falls just under the generic 0.01 BS1 threshold.
BS2 Not met: three homozygous carriers (gnomAD v4.1, gnomAD-Canada) but POLE disease evaluated here is adult-onset and incompletely penetrant, failing BS2's early-onset premise.
BS3 Not assessed: no assay demonstrates preserved POLE splicing or proofreading; SpliceAI max delta 0.003 is in-silico only, not functional evidence.
BS4 Not assessed: no pedigree with affected relatives and genotypes exists to assess non-segregation.
BP2 Not assessed: no phase or companion-variant data exist to place this allele in cis or trans with a pathogenic POLE variant.
BP5 Not assessed: the case contains no proband genotype or molecular diagnosis, so BP5's alternate-molecular-basis requirement cannot be evaluated.
BP6 Not met: 0 expert-panel ClinVar submissions for this variant, so the single-submitter Likely benign label cannot satisfy BP6.
N/A · 7 PS1 · PM4 · PM5 · PP2 · BP1 · BP3 · BP7
Research & evidence
Population frequency
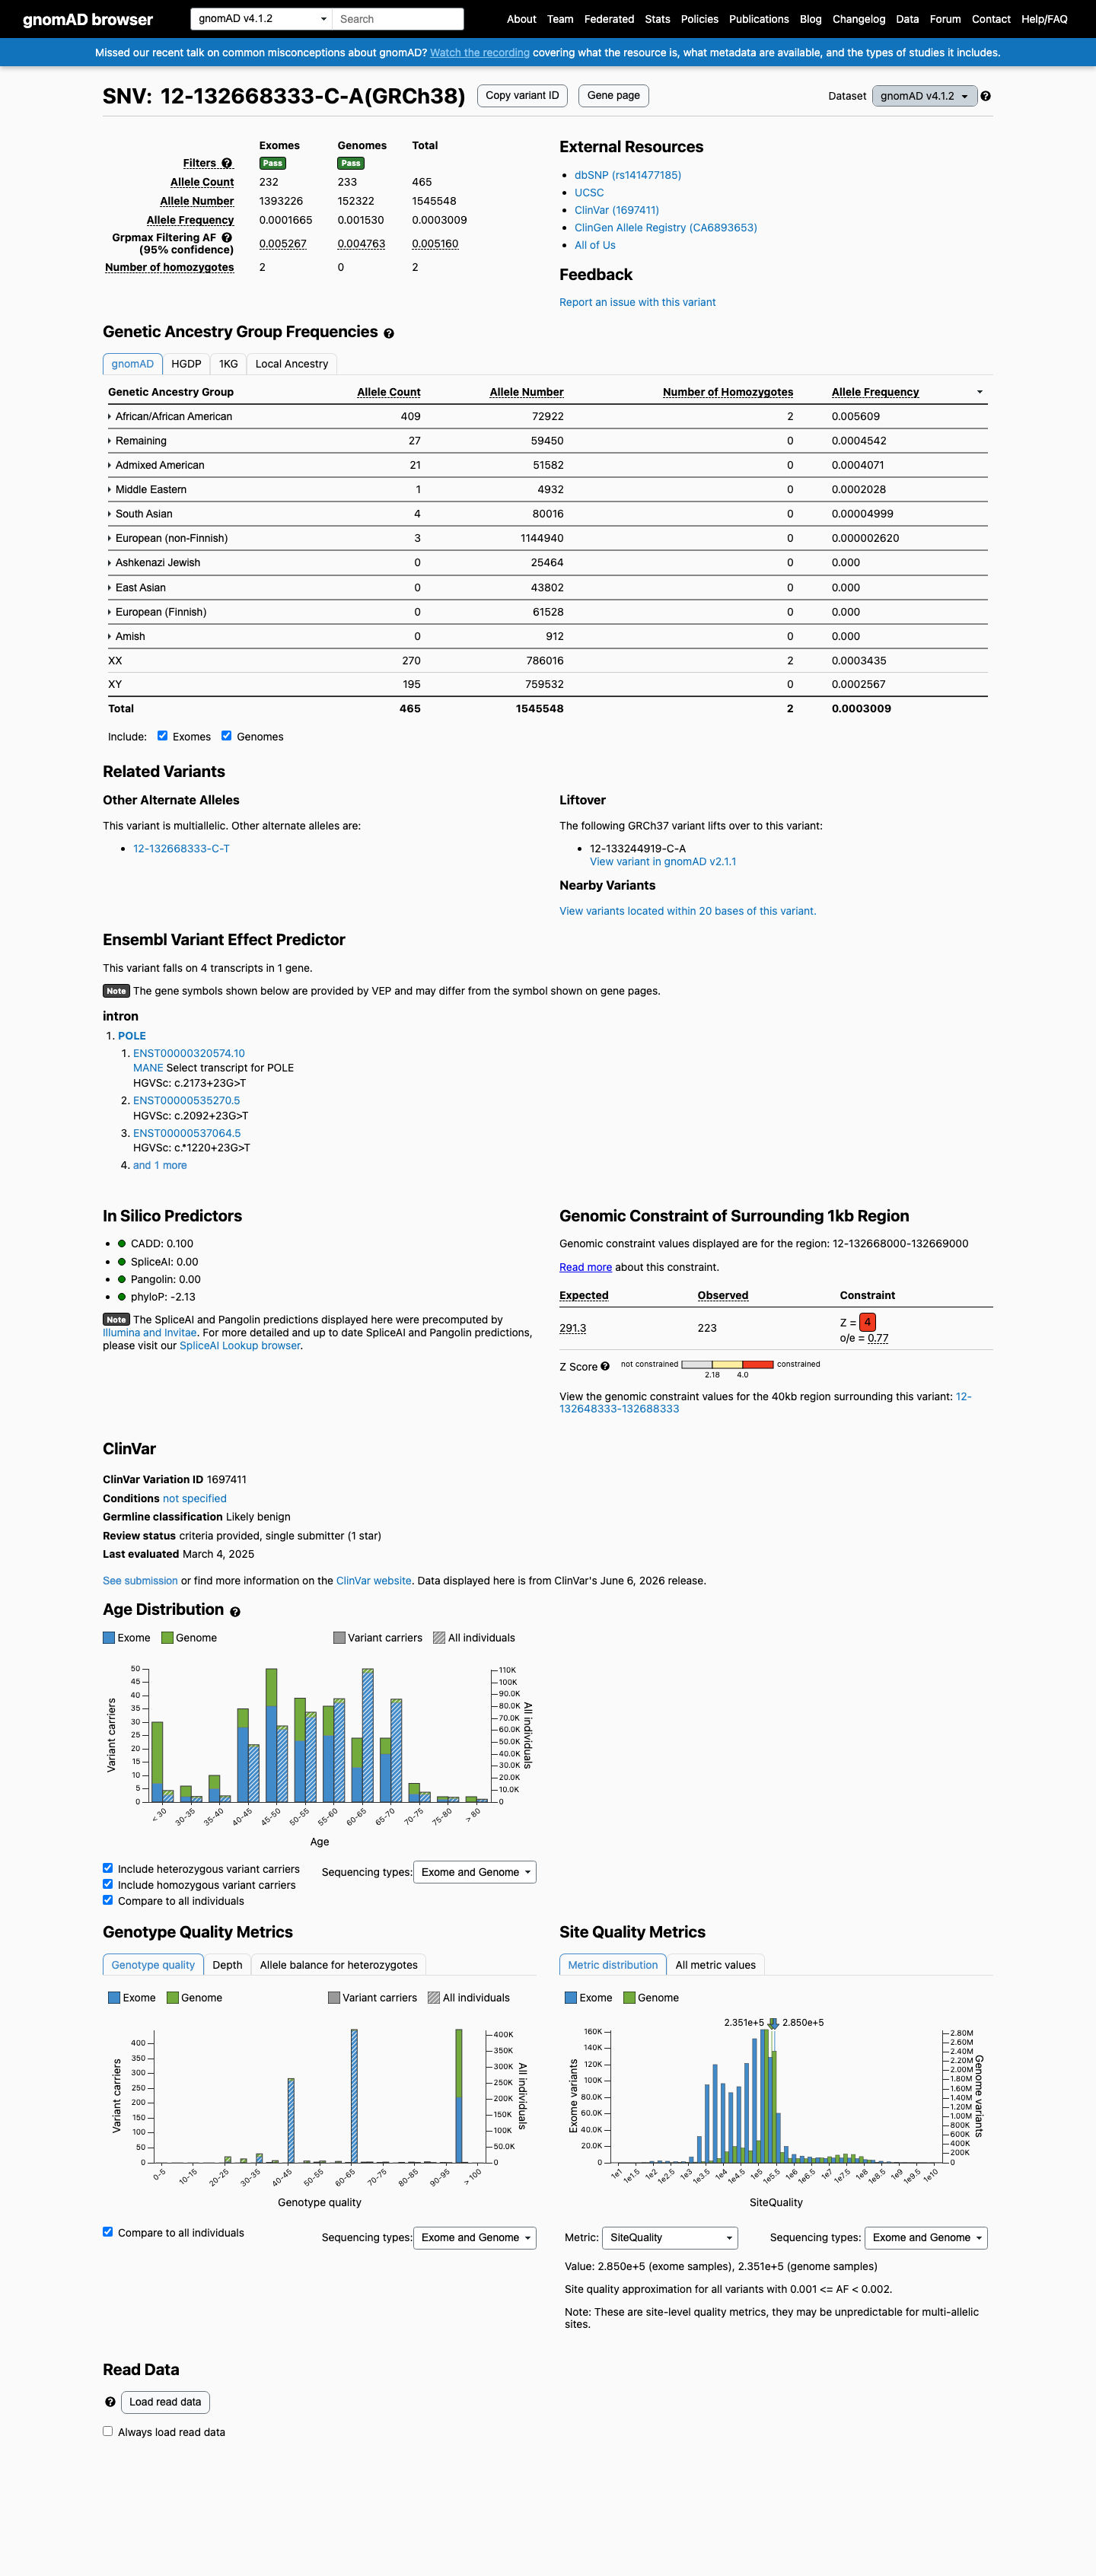
gnomAD v4.1 screenshot
gnomAD v4.1
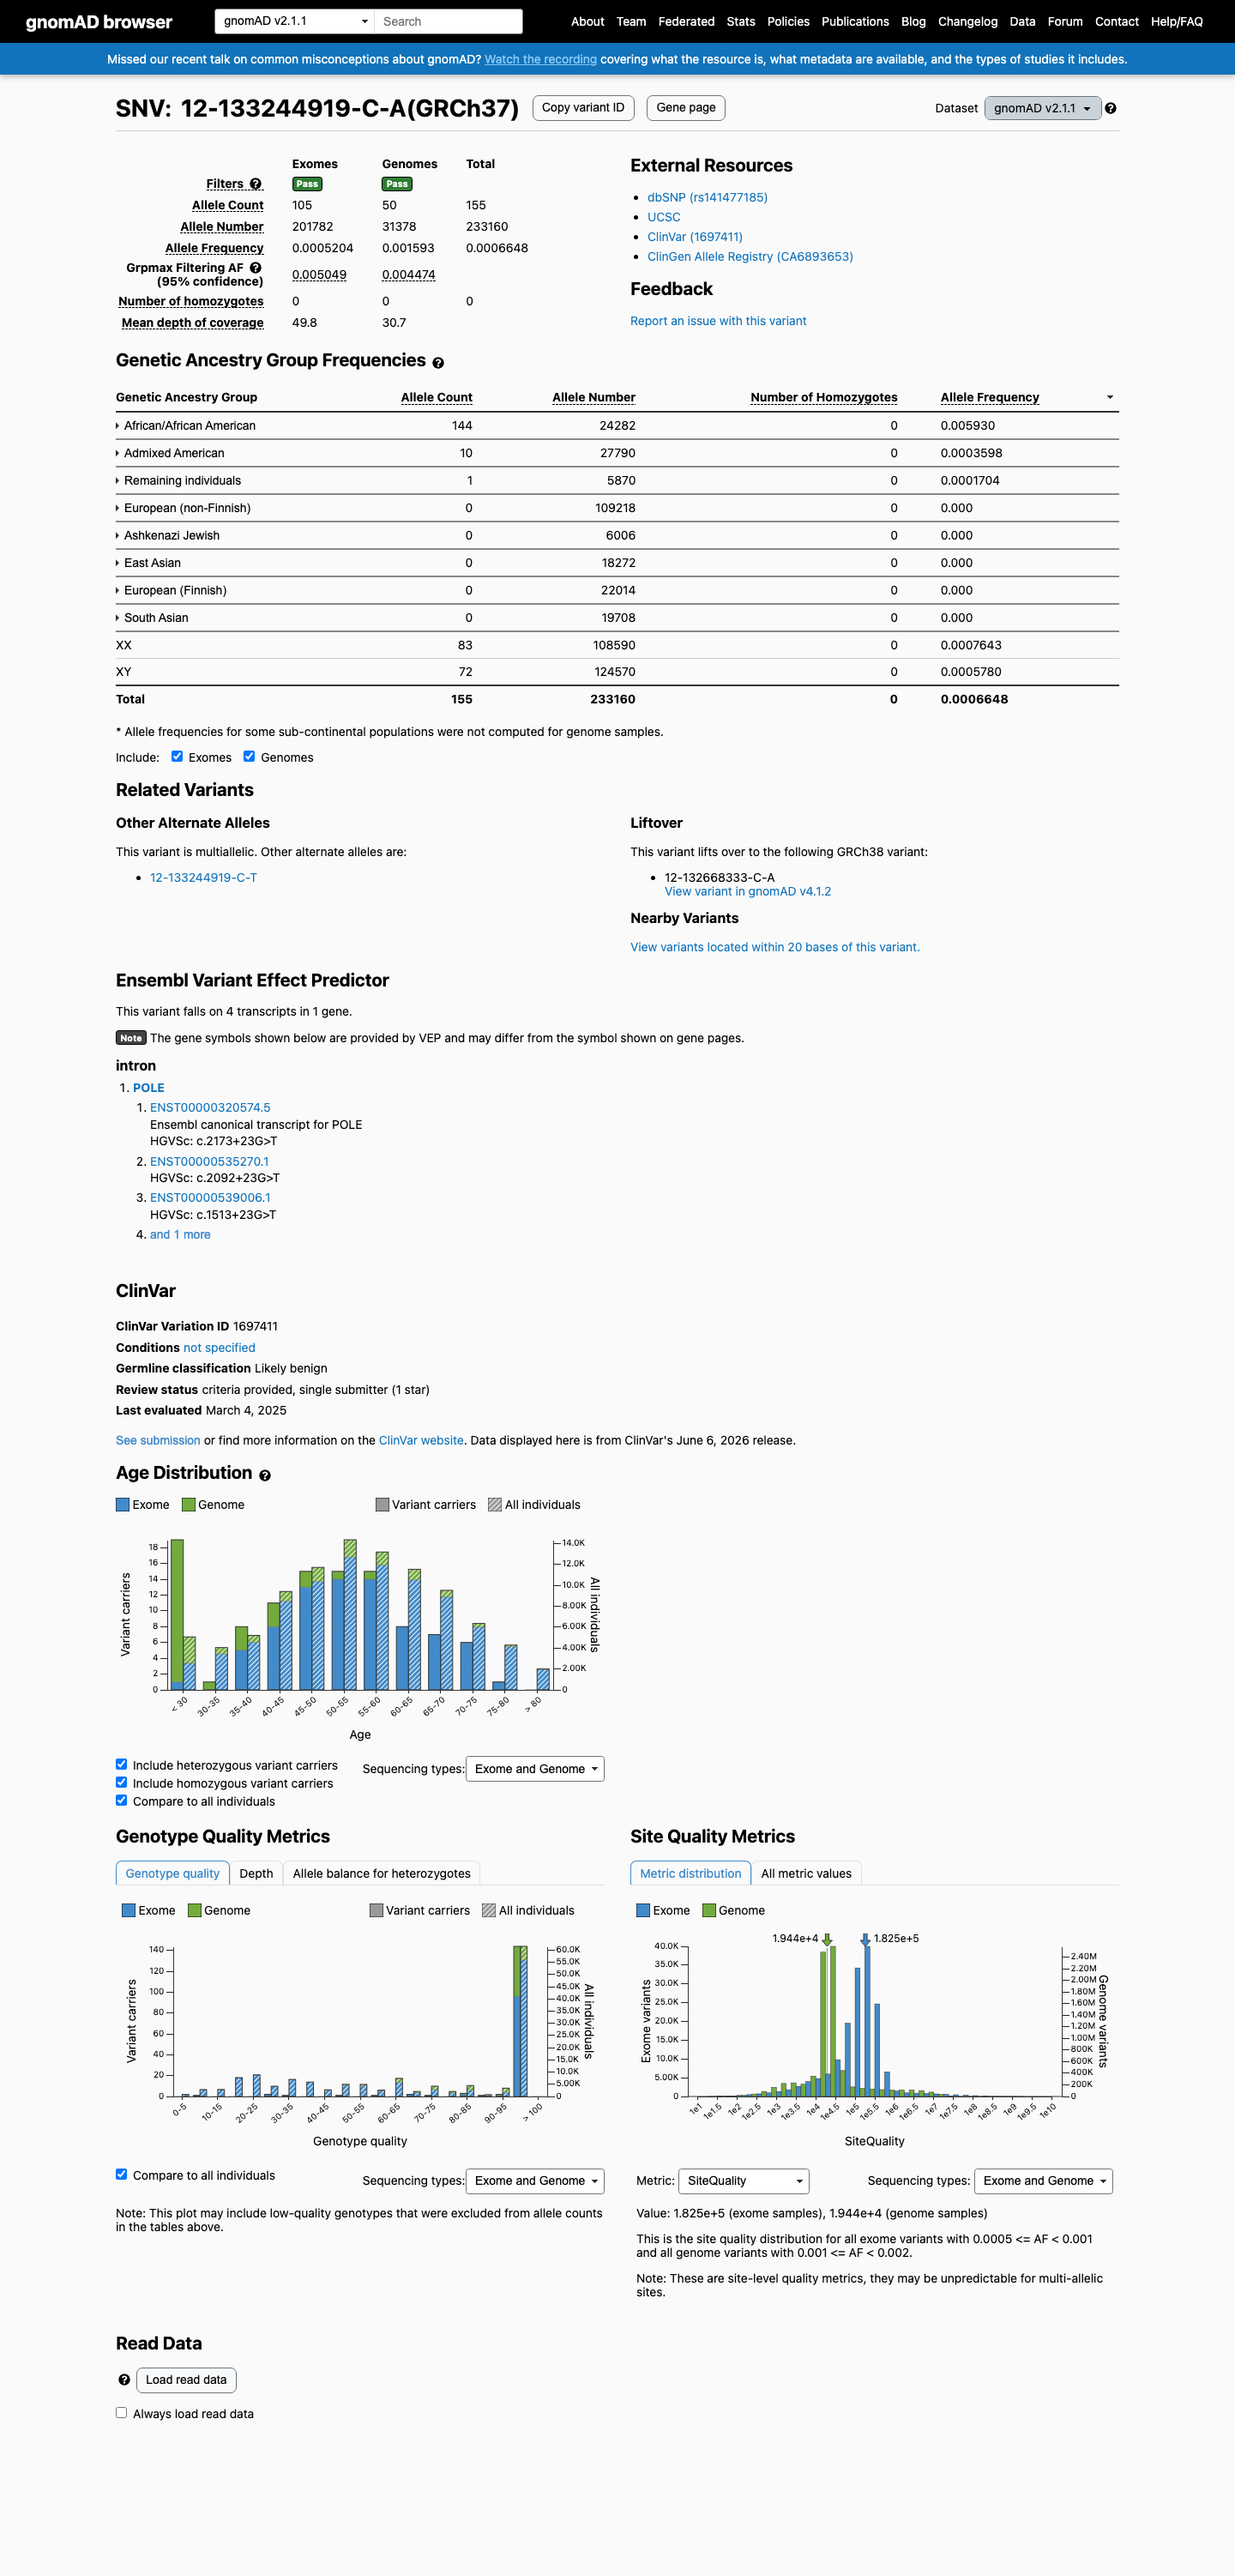
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 0.000300864; MAF= 0.03009%, 465/1545548 alleles, homozygotes = 2) and has highest observed frequency in the African/African American population (AF= 0.00560873; MAF= 0.56087%, 409/72922 alleles, homozygotes = 2); grpmax FAF= 0.00515979.
v2.1
This variant is present in gnomAD v2.1 (AF= 0.00066478; MAF= 0.06648%, 155/233160 alleles, homozygotes = 0) and has highest observed frequency in the African/African American population (AF= 0.00593032; MAF= 0.59303%, 144/24282 alleles, homozygotes = 0); grpmax FAF= 0.00504945.
🇨🇦 CA
This variant is present in gnomAD-Canada v1.0 (AF= 0.0005428881650380022, 10/18420 alleles, homozygotes = 1).
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.03% · 465 / 1,545,548
2 hom · FAF 0.52%
African/African American
409 / 72,922
0.56%
2 hom
Remaining individuals
27 / 59,450
0.045%
Admixed American
21 / 51,582
0.041%
Middle Eastern
1 / 4,932
0.02%
South Asian
4 / 80,016
0.005%
European (non-Finnish)
3 / 1,144,940
0.00026%
+ 4 not observed (European (Finnish), Amish, East Asian, Ashkenazi Jewish)
gnomAD v2.1
0.066% · 155 / 233,160
0 hom · FAF 0.5%
African/African American
144 / 24,282
0.59%
Admixed American
10 / 27,790
0.036%
Remaining individuals
1 / 5,870
0.017%
+ 5 not observed (Ashkenazi Jewish, East Asian, European (Finnish), European (non-Finnish), South Asian)
gnomAD Canada 🇨🇦
0.054% · 10 / 18,420
1 hom · FAF 0.53%
indel · split
African/African American
10 / 1,020
0.98%
1 hom
+ 8 not observed (Latino/Admixed American, Ashkenazi Jewish, East Asian, European (Finnish), Middle Eastern, European (non-Finnish), Remaining individuals, South Asian)
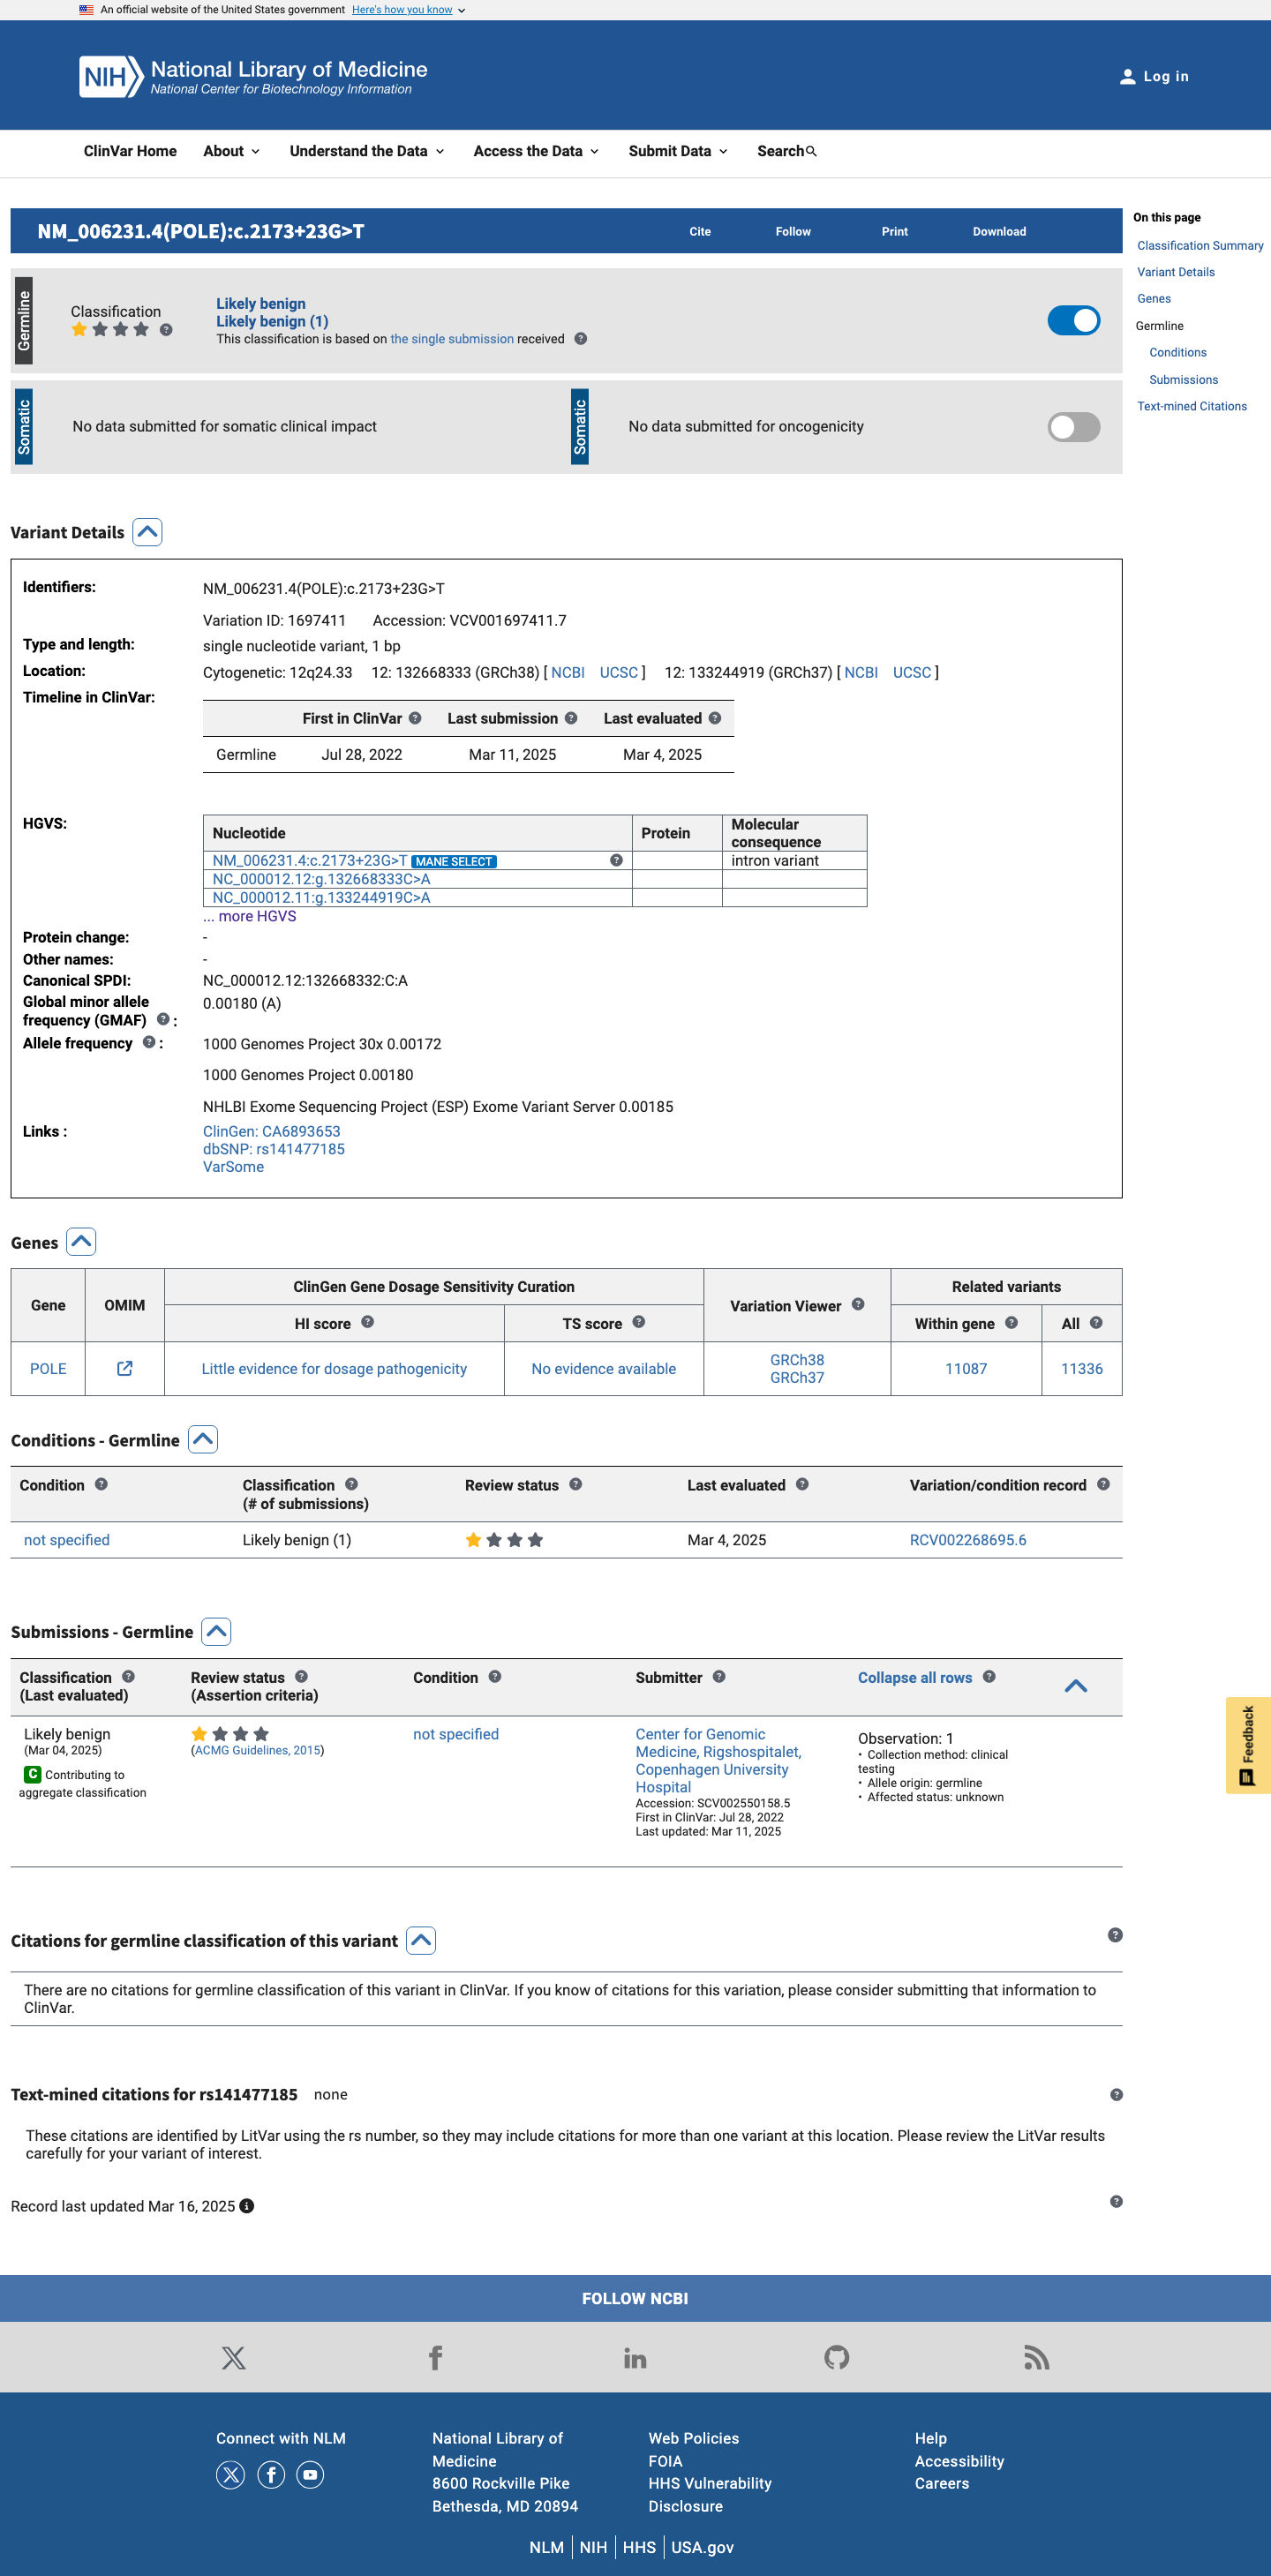
ClinVar screenshot
ClinVar
This variant is present in ClinVar (Variation ID: 1697411); submission details unavailable.
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.00).
Functional No data
No calibrated functional assay or RNA evidence was identified for this variant.
OncoKB ↗
COSMIC screenshot
COSMIC
Somatic evidence
COSMIC
This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant cancer hotspot.
COSMIC ↗
Sources & reference links
7Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC