Back
NM_006231.4:c.4039C>T
p.Leu1347= · POLE
León-Castillo et al. 2020 custom POLE framework · vleon-castillo-2020-custom-framework-v1
0%
complete
Final classification
VUS
PM2BP7
POLE
c.4039C>T
p.Leu1347=
synonymous · exon 32

POLE encodes the catalytic subunit of DNA polymerase epsilon, the enzyme that replicates the leading strand of DNA during cell division and participates in DNA repair. It contains a proofreading domain that corrects replication errors, keeping the accumulation of mutations in check. Germline mutations in POLE cause polyposis and predispose to colorectal cancer, and are also linked to a rare syndrome of facial dysmorphism, immunodeficiency, livedo, and short stature. Somatic mutations, particularly in the proofreading domain, occur in colorectal and endometrial cancers, where they drive an ultra-mutated tumor phenotype and are associated with better responses to immune checkpoint inhibitors.

This variant

POLE encodes the leading-strand DNA polymerase epsilon proofreading subunit, and the synonymous change p.(Leu1347=) leaves that proofreading protein unchanged while sitting outside the exonuclease domain in which the established colorectal and endometrial cancer predisposition variants are concentrated.

Transcript
NM_006231.4
HGVS · transcript:coding
NM_006231.4:c.4039C>T
GRCh38
chr12:132649039 G>A
GRCh37
chr12:133225625 G>A
VUS: PM2 (supporting) plus BP7 (supporting) meet opposing ACMG/AMP thresholds, matching no Pathogenic, Likely Pathogenic, Benign, or Likely Benign rule under the local POLE framework.
Classification rationale
PM2 BP7 VUS
POLE c.4039C>T synonymous · exon 32

VUS: PM2 (supporting) is met because the gnomAD v4.1 all-comers allele frequency of 1.86e-06 is far below the 1e-4 rarity threshold. VUS: BP7 (supporting) is met because this synonymous variant's SpliceAI maximum delta of 0.014 predicts no splicing impact. VUS: one supporting pathogenic plus one supporting benign criterion is a conflicting combination that satisfies no classification rule.

PM2 + BP7 → VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_006231.4 · variants mapped to exon structure
POLE NM_006231.4
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 19 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
PM2 supporting Pathogenic
Met (supporting): gnomAD v4.1 frequency 1.86e-06 and group-maximum 2.80e-07 both fall below the 1e-4 PM2 threshold.
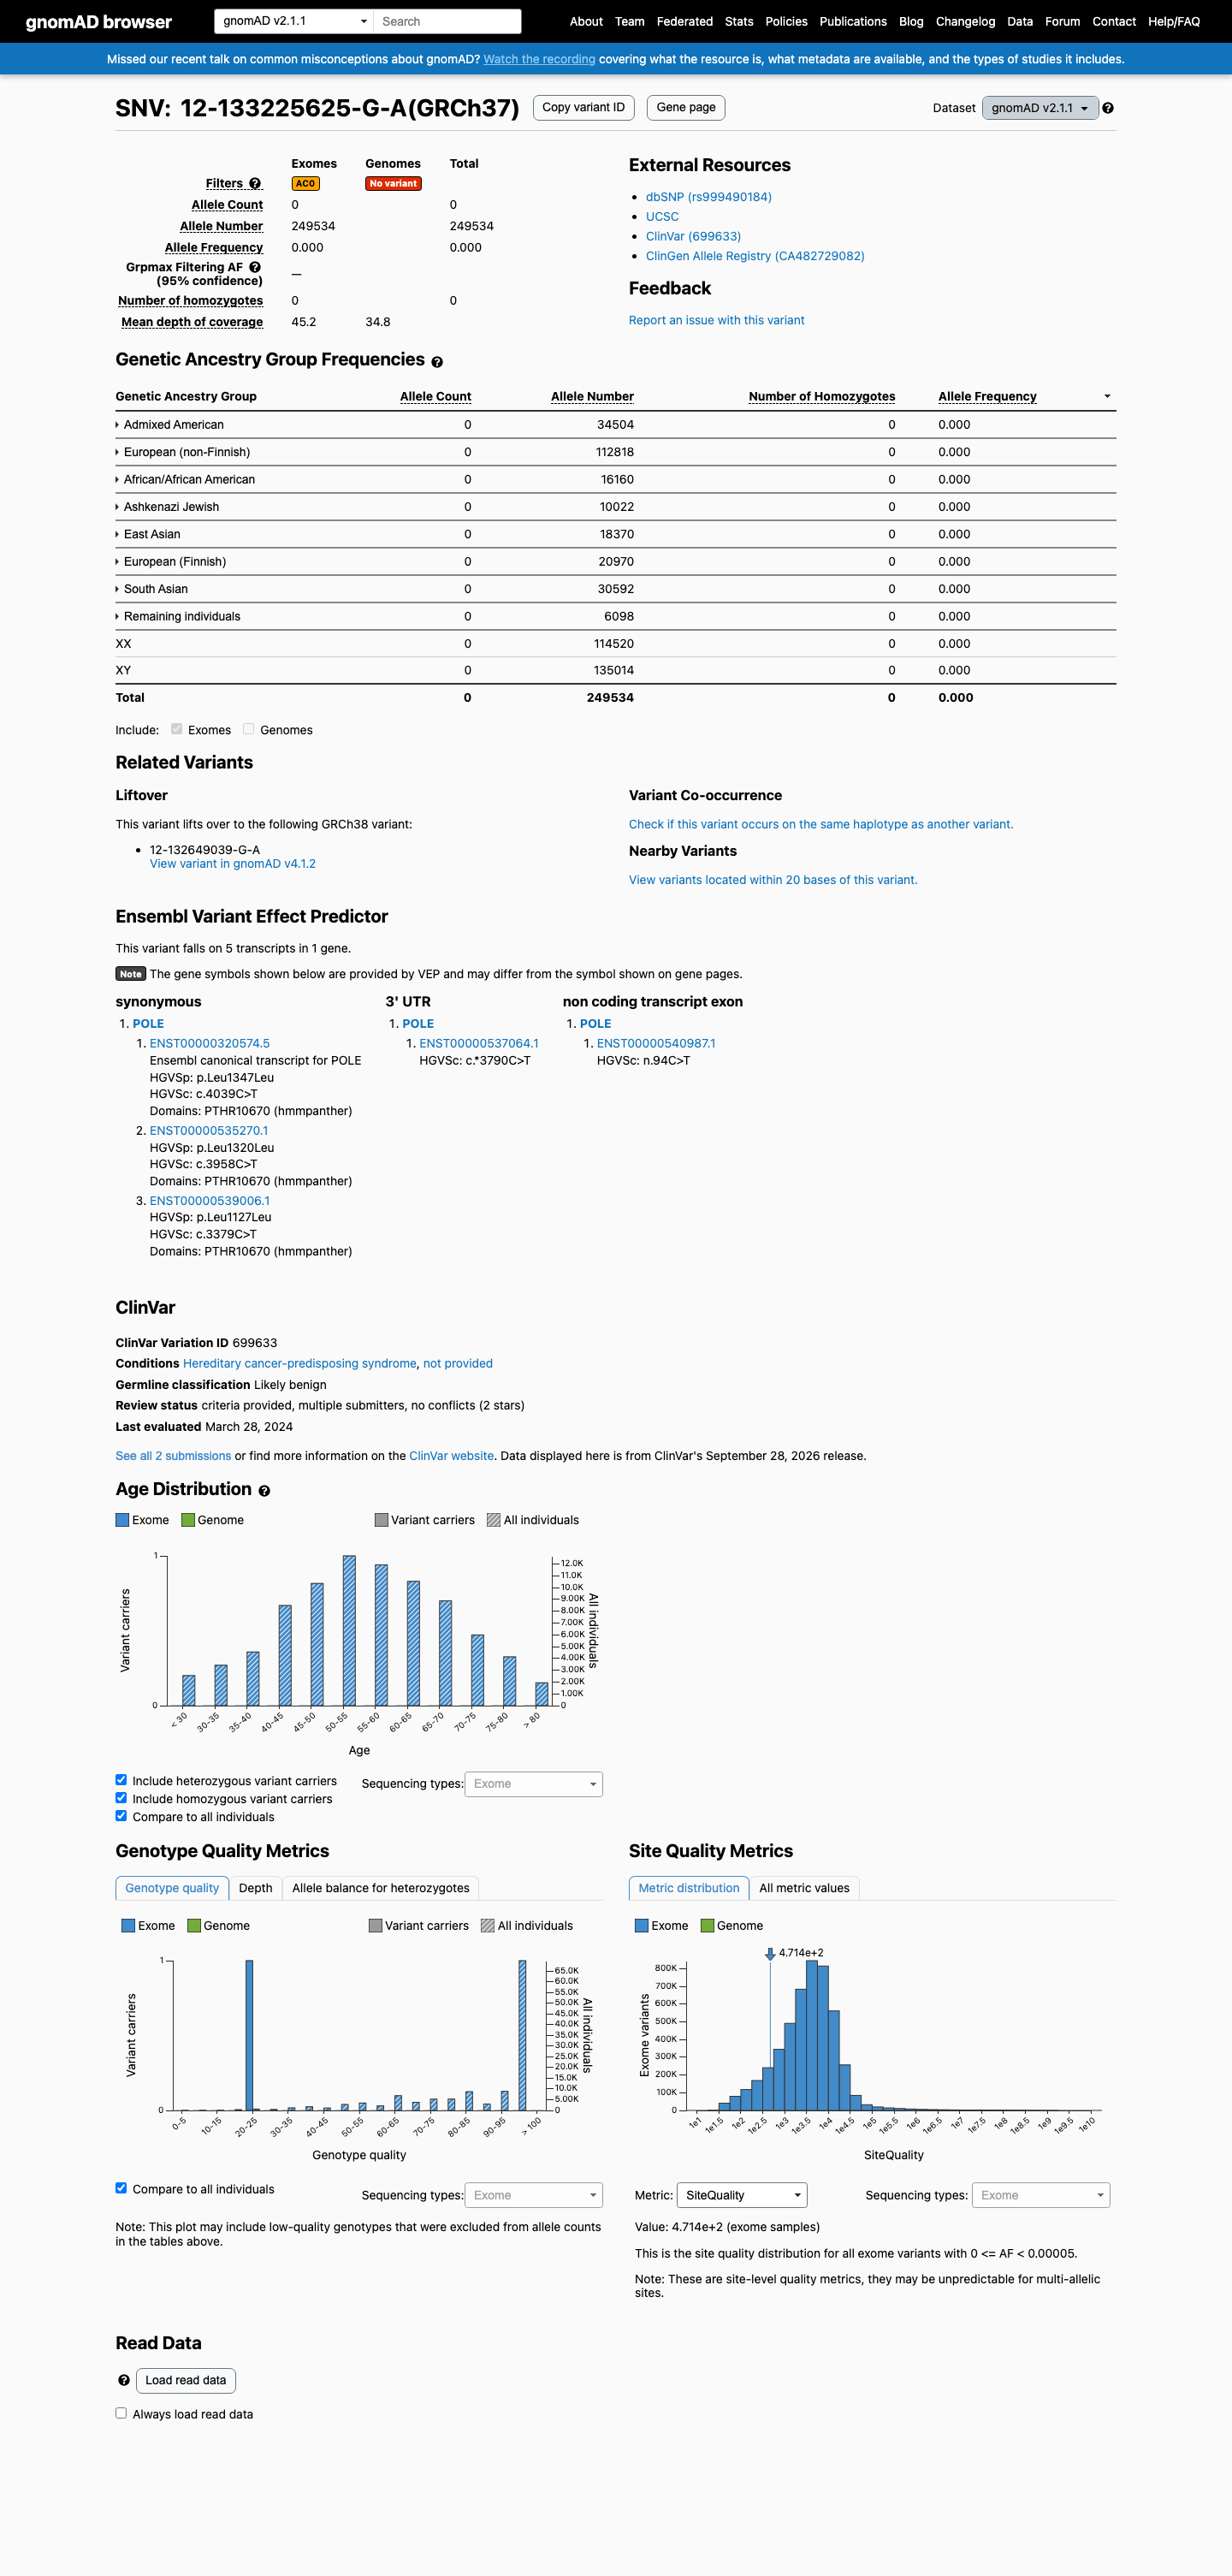
gnomAD v4.1 (all-comers): total AF 1.85954e-06 (3/1613304 alleles); group-maximum filtering AF 2.8e-07; highest subpopulation AF 1.09801e-05 (1/91074, South Asian); zero homozygotes.gnomAD v2.1 (all-comers, exome): variant absent, 0/249534 alleles, AF 0.gnomAD v2.1 non-cancer exomes (0/235300) and gnomAD v3.1 non-cancer genomes (absent) are concordant and equally support PM2, but are not substituted because no governing POLE specification requires a non-cancer or exome-only source.
BP7 supporting Benign
Met, supporting: synonymous variant with SpliceAI max delta 0.014, far below the <=0.1 no-splice-impact threshold.
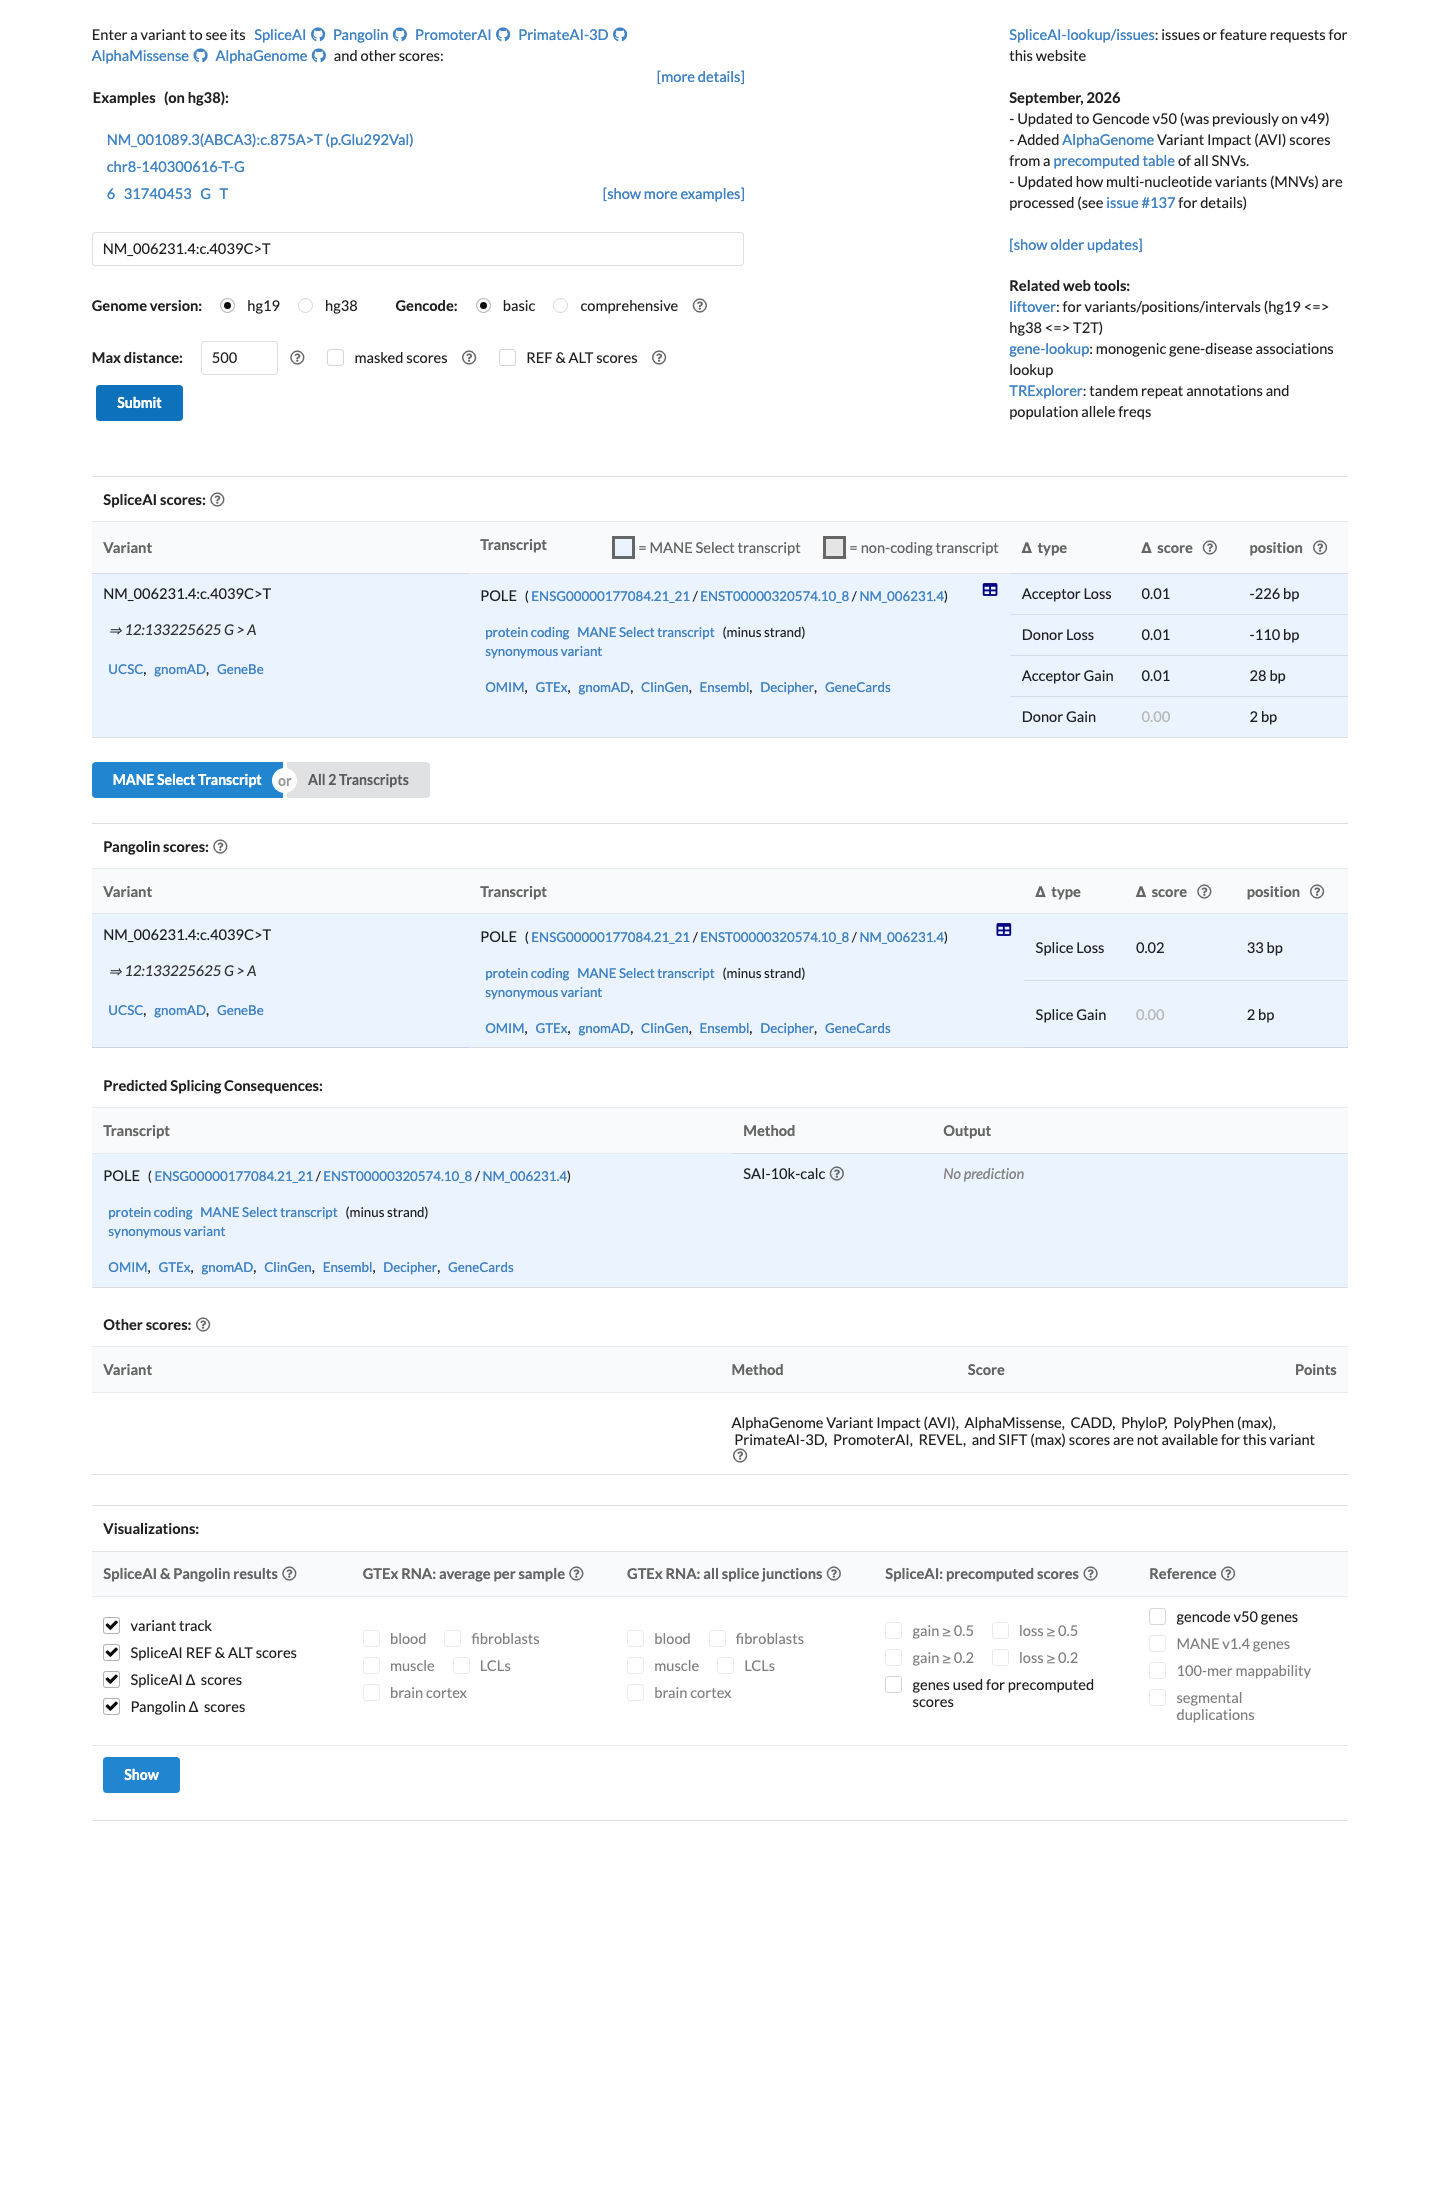
The normalized consequence is synonymous: NM_006231.4:c.4039C>T produces NP_006222.2:p.(Leu1347=) (protein_1l p.(L1347=)), and the variant-level PVS1 scaffold records consequence_class 'synonymous' with canonical_splice_consensus false.SpliceAI Lookup result for NM_006231.4:c.4039C>T (12-133225625-G-A, hg37): DS_AG 0.014, DS_AL 0.008, DS_DG 0.001, DS_DL 0.01; maximum delta score 0.014; Pangolin SG 0.003, SL -0.017 - no native acceptor/donor site lost and no new splice site created.Published SpliceAI calibration used verbatim: BP4/no-significant-impact cutoff at max delta <=0.1 (Jaganathan et al. 2019, Cell, PMID:30661751); the observed 0.014 is well below it, applied here as the calibrated splice-impact basis for BP7.
Assessed · not applied · 11 not met · 8 not assessed
Pathogenic
PS2 Not assessed: no proband, parental testing, parentage confirmation, or family-history data exist for this variant in the case evidence.
PS3 Not met: no functional RNA/splice or protein assay exists for p.(Leu1347=); SpliceAI max delta 0.014 is in-silico and cannot support PS3.
PS4 Not met: this synonymous variant is absent from Supplementary Table S1, so its recurrence count of 0 fails the framework's combined EC count >=10 requirement.
PM1 Not met: p.Leu1347= is not one of the framework's enumerated exonuclease-domain hotspot substitutions and lies outside the exonuclease domain, whose hotspots span residues 286-459.
PM3 Not assessed: no trans/phase data - no second POLE variant or parental genotypes - so PM3's requirement for a pathogenic variant in trans cannot be evaluated.
PM6 Not assessed: no proband or parent-of-origin data exist, so no assumed de novo occurrence can be documented for this variant.
PP1 Not assessed: no pedigree, affected relatives, or meioses are available, so co-segregation of c.4039C>T with disease cannot be evaluated.
PP3 Not met: synonymous variant with SpliceAI max delta 0.014, far below the >=0.2 PP3 supporting threshold.
PP4 Not assessed: no proband phenotype or family-history data was recorded for this case, so phenotype specificity for a single-gene disorder could not be evaluated.
PP5 Not met: ClinVar classifies this variant Likely benign from 2 single-submitter laboratories with 0 expert-panel submissions, failing the expert-panel pathogenic requirement.
Benign
BA1 Not met: the highest gnomAD frequency is 1.10e-05 (South Asian), far below the 0.05 BA1 stand-alone benign threshold.
BS1 Not met: the highest gnomAD frequency, 1.10e-05, is well below the 0.01 BS1 strong benign threshold.
BS2 Not met: zero homozygotes and only 3 carrier alleles in gnomAD, with no unaffected-adult evidence required by BS2.
BS3 Not met: no experimental assay shows a non-damaging effect for p.(Leu1347=); SpliceAI max delta 0.014 is in-silico, and ClinVar status is not a functional dataset.
BS4 Not assessed: no family with genotyped affected and unaffected members exists, so non-segregation of c.4039C>T cannot be demonstrated.
BP2 Not assessed: no co-occurring pathogenic variant and no phasing data, so BP2's in-trans-or-in-cis requirement for c.4039C>T is unevidenced.
BP4 Not met: the single SpliceAI max delta 0.014 supports no splice impact but is allocated to BP7, leaving no independent BP4 evidence.
BP5 Not assessed: no case-level genotype or co-occurring-variant data was available, so an alternate molecular basis for disease could not be evaluated.
BP6 Not met: the ClinVar Likely benign classification carries 0 expert-panel submissions, so BP6's expert-panel benign-source requirement is not satisfied.
N/A · 7 PVS1 · PS1 · PM4 · PM5 · PP2 · BP1 · BP3
Research & evidence
Population frequency
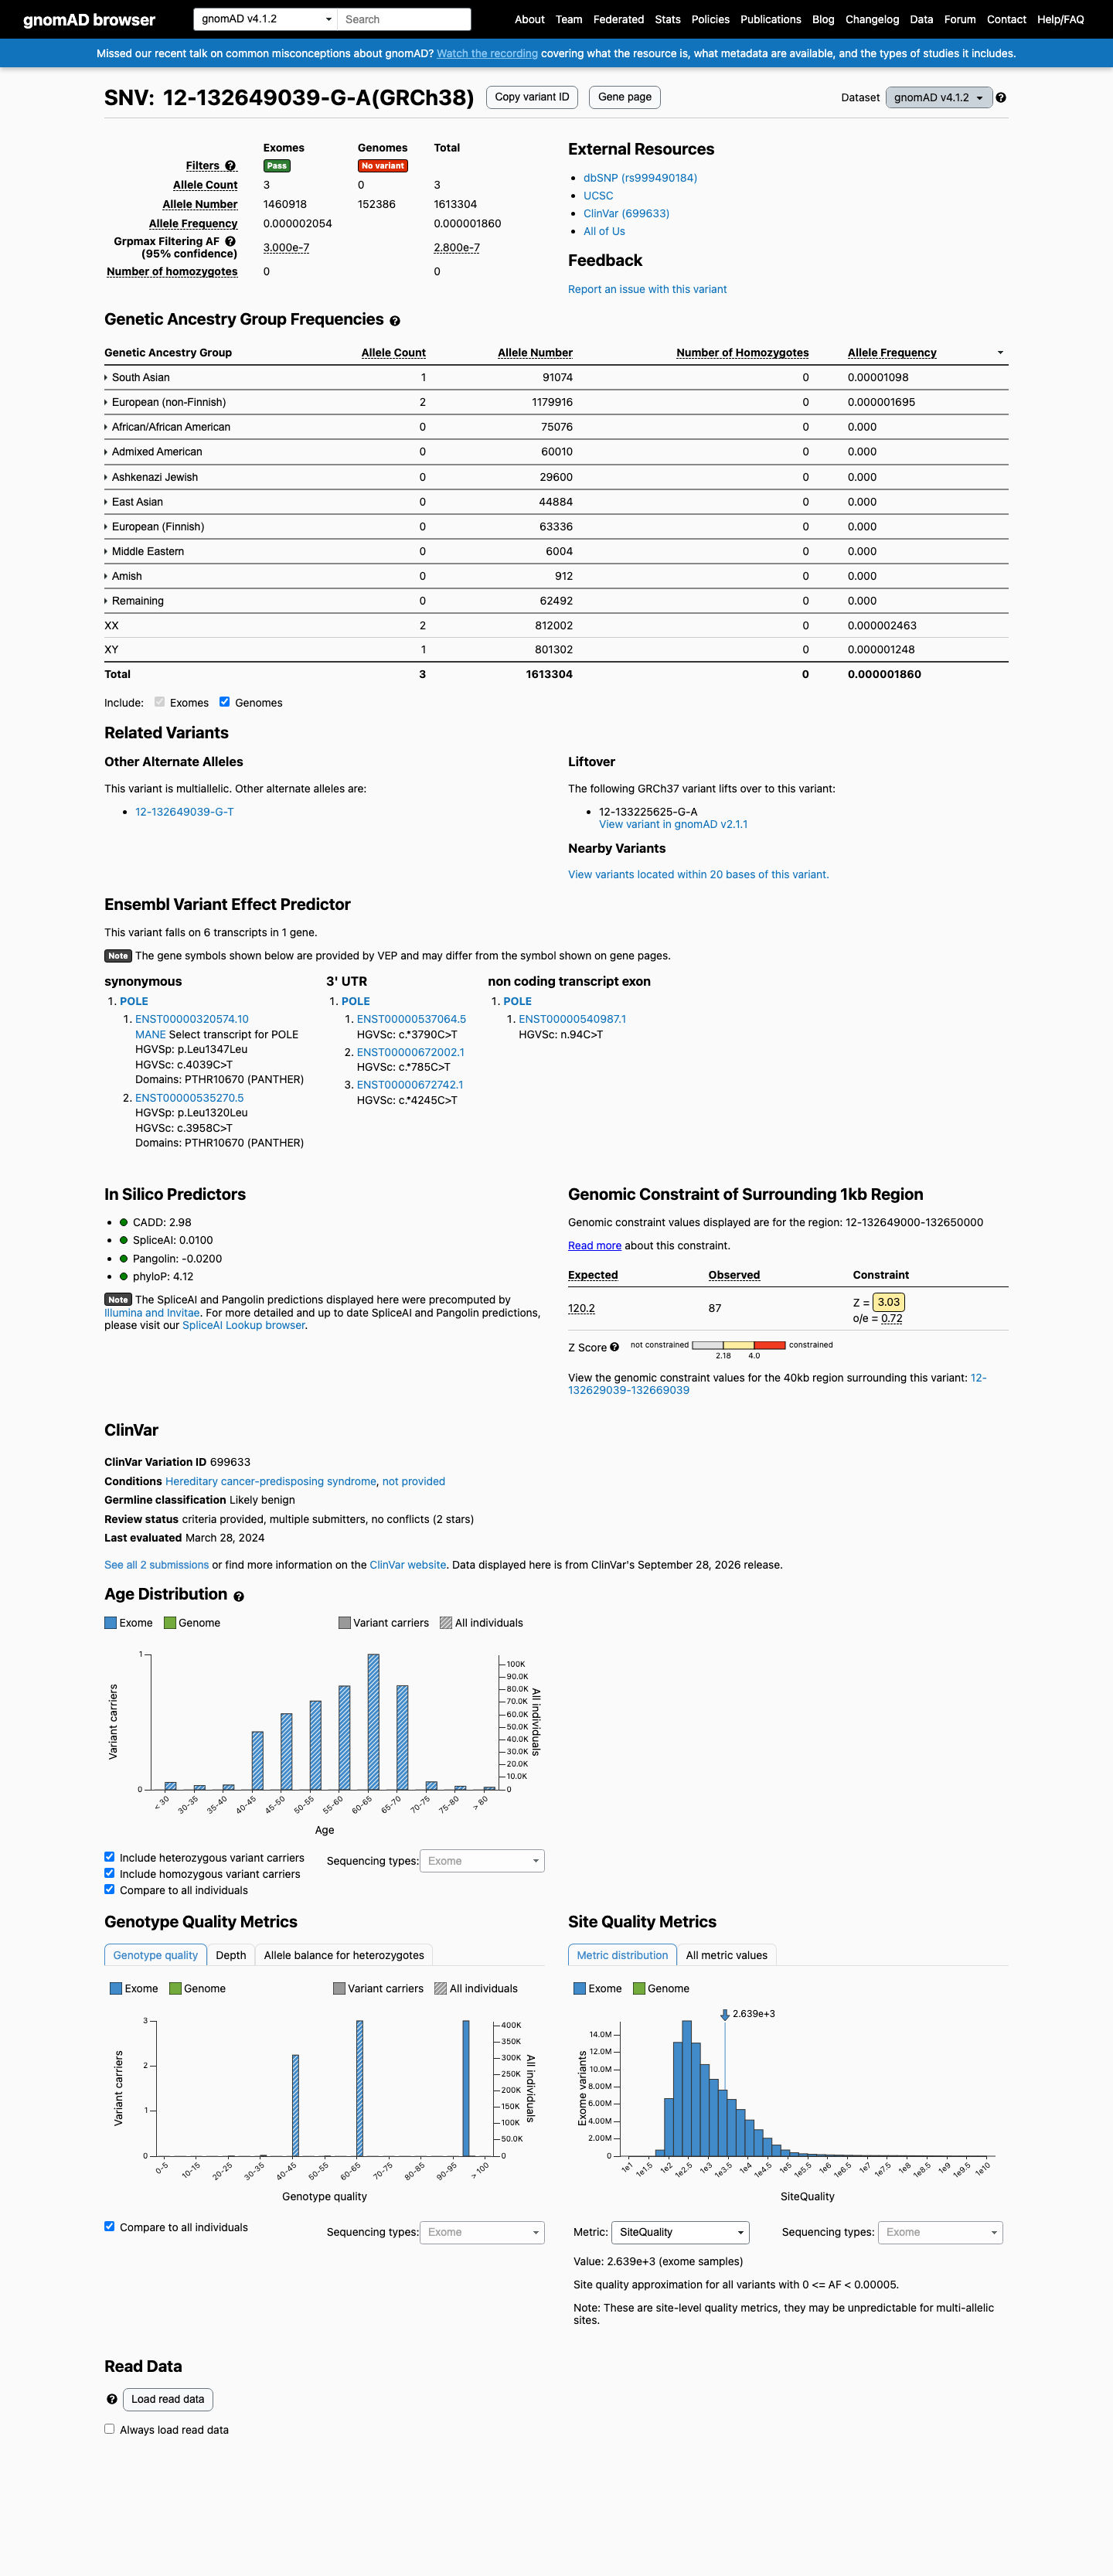
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 1.85954e-06; MAF= 0.00019%, 3/1613304 alleles, homozygotes = 0) and has highest observed frequency in the South Asian population (AF= 1.09801e-05; MAF= 0.00110%, 1/91074 alleles, homozygotes = 0); grpmax FAF= 2.8e-07.
v2.1
This variant is present in gnomAD v2.1 (AF= 0; MAF= 0.00000%, 0/249534 alleles, homozygotes = 0) and has highest observed frequency in the African/African American population (AF= 0; MAF= 0.00000%, 0/16160 alleles, homozygotes = 0).
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.00019% · 3 / 1,613,304
0 hom · FAF 2.8e-05%
South Asian
1 / 91,074
0.0011%
European (non-Finnish)
2 / 1,179,916
0.00017%
+ 8 not observed (Remaining individuals, Admixed American, European (Finnish), Amish, East Asian, Middle Eastern, Ashkenazi Jewish, African/African American)
gnomAD v2.1
Absent · 0 / 249,534
0 hom
Not observed in any ancestry group.
+ 8 not observed (African/African American, Admixed American, Ashkenazi Jewish, East Asian, European (Finnish), European (non-Finnish), Remaining individuals, South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
ClinVar screenshot
ClinVar
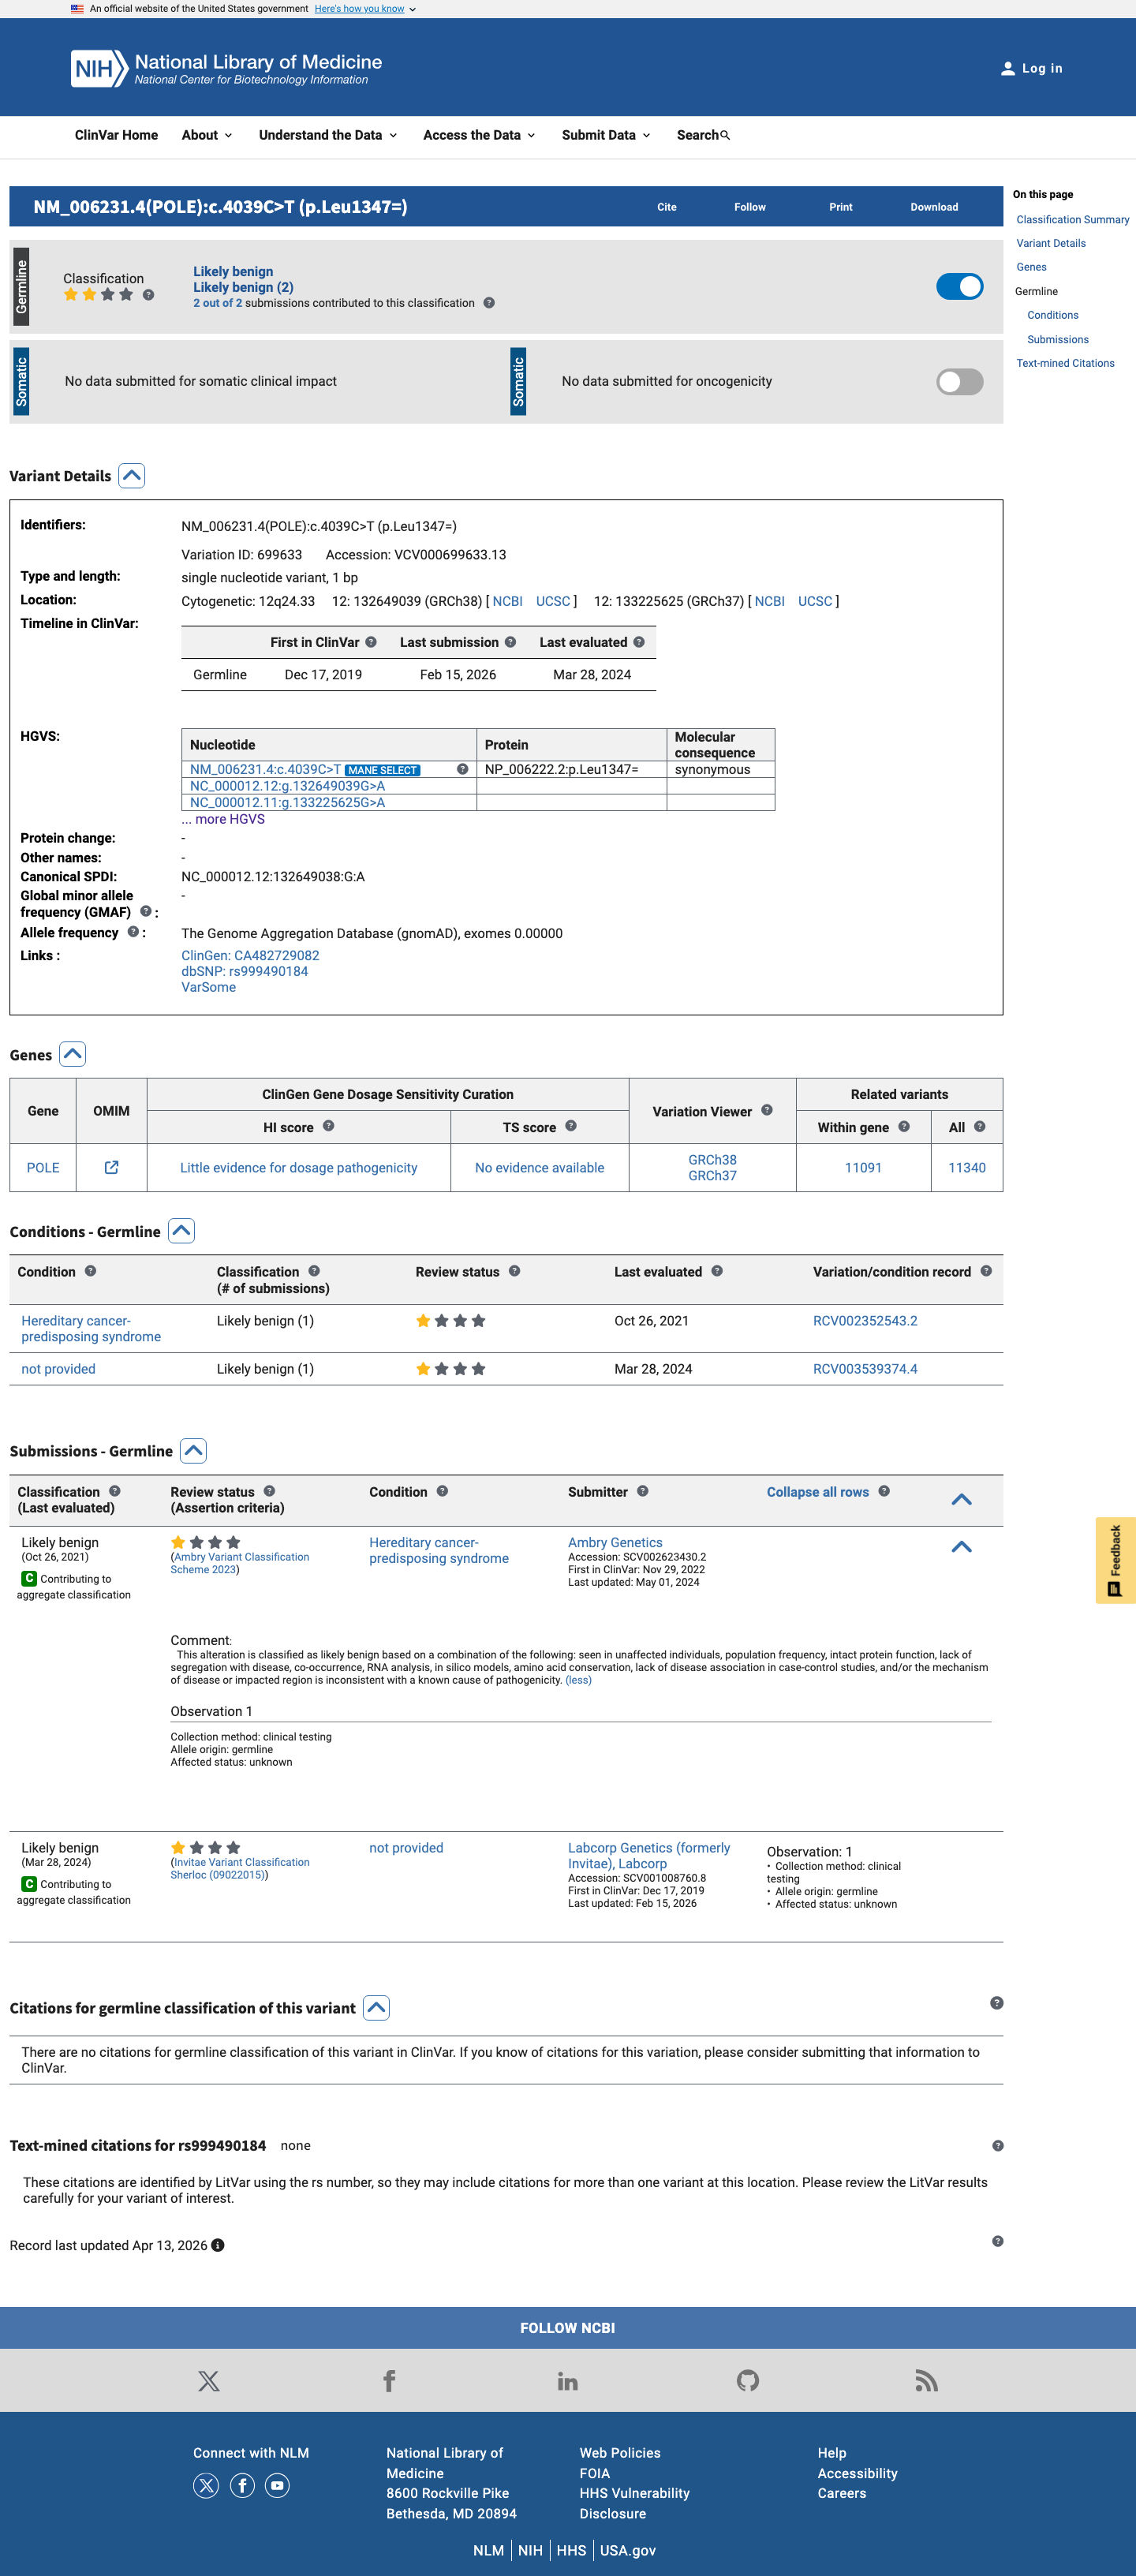
This variant has been reported in ClinVar as Likely benign (2 clinical laboratories). (ClinVarID = 699633)
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.01).
Functional / OncoKB screenshot
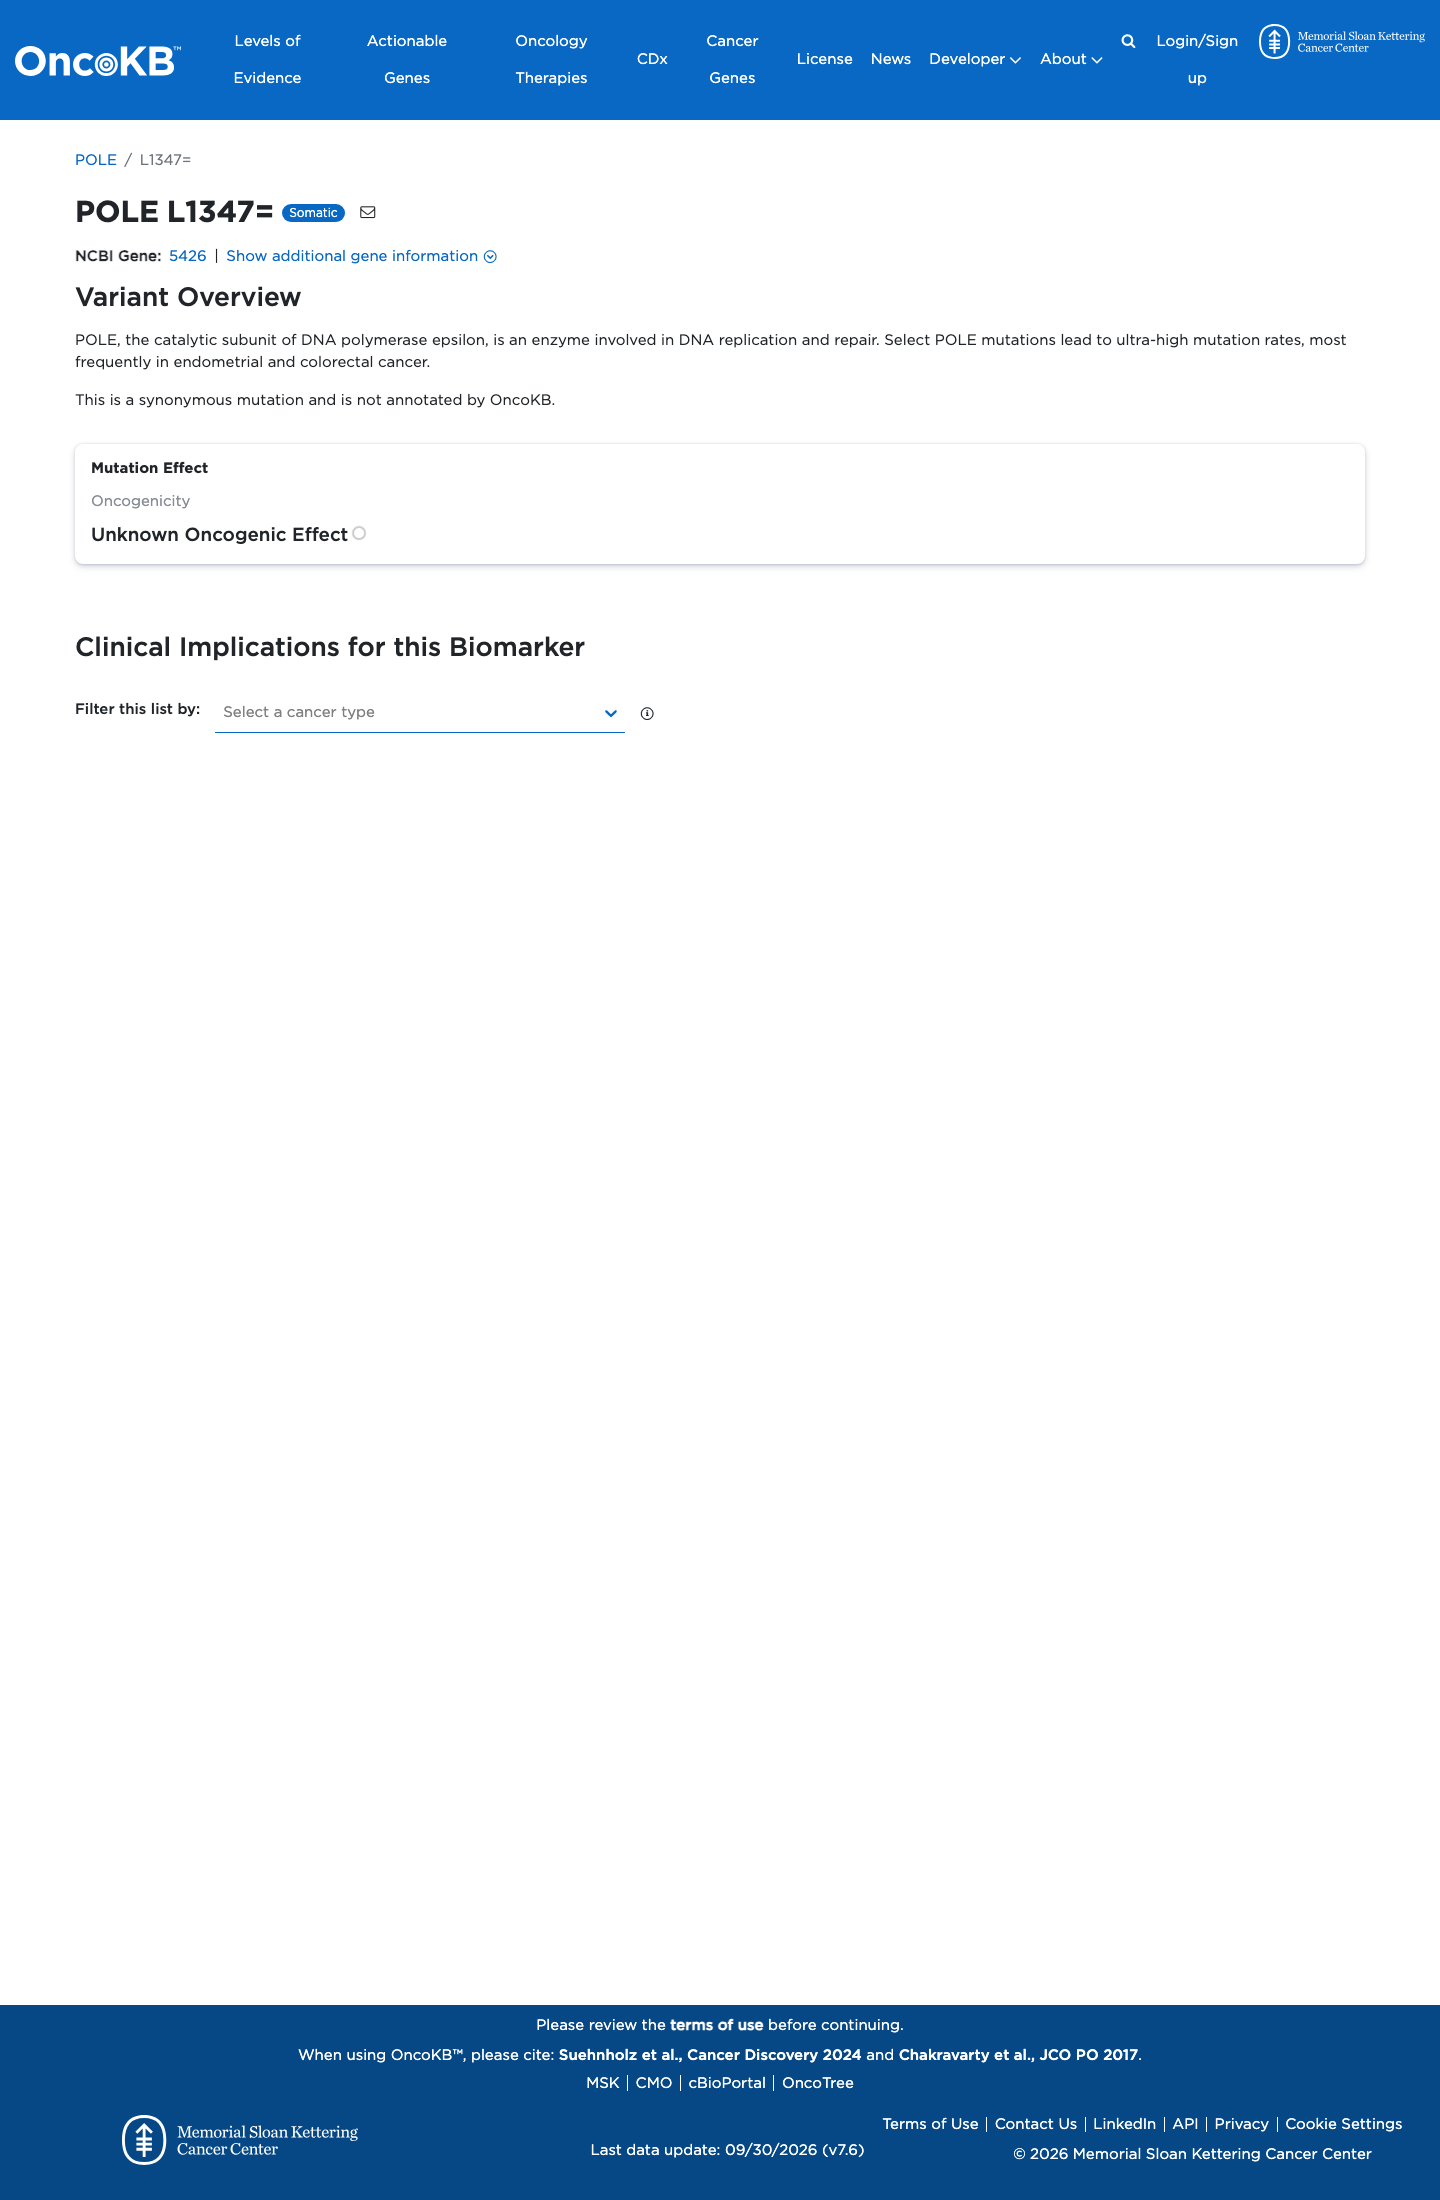
Functional Unknown Oncogenic Effect
OncoKB identified curated literature and non-variant-specific oncogenicity context for review; listed oncogenicity label: Unknown Oncogenic Effect.
OncoKB ↗
COSMIC screenshot
COSMIC
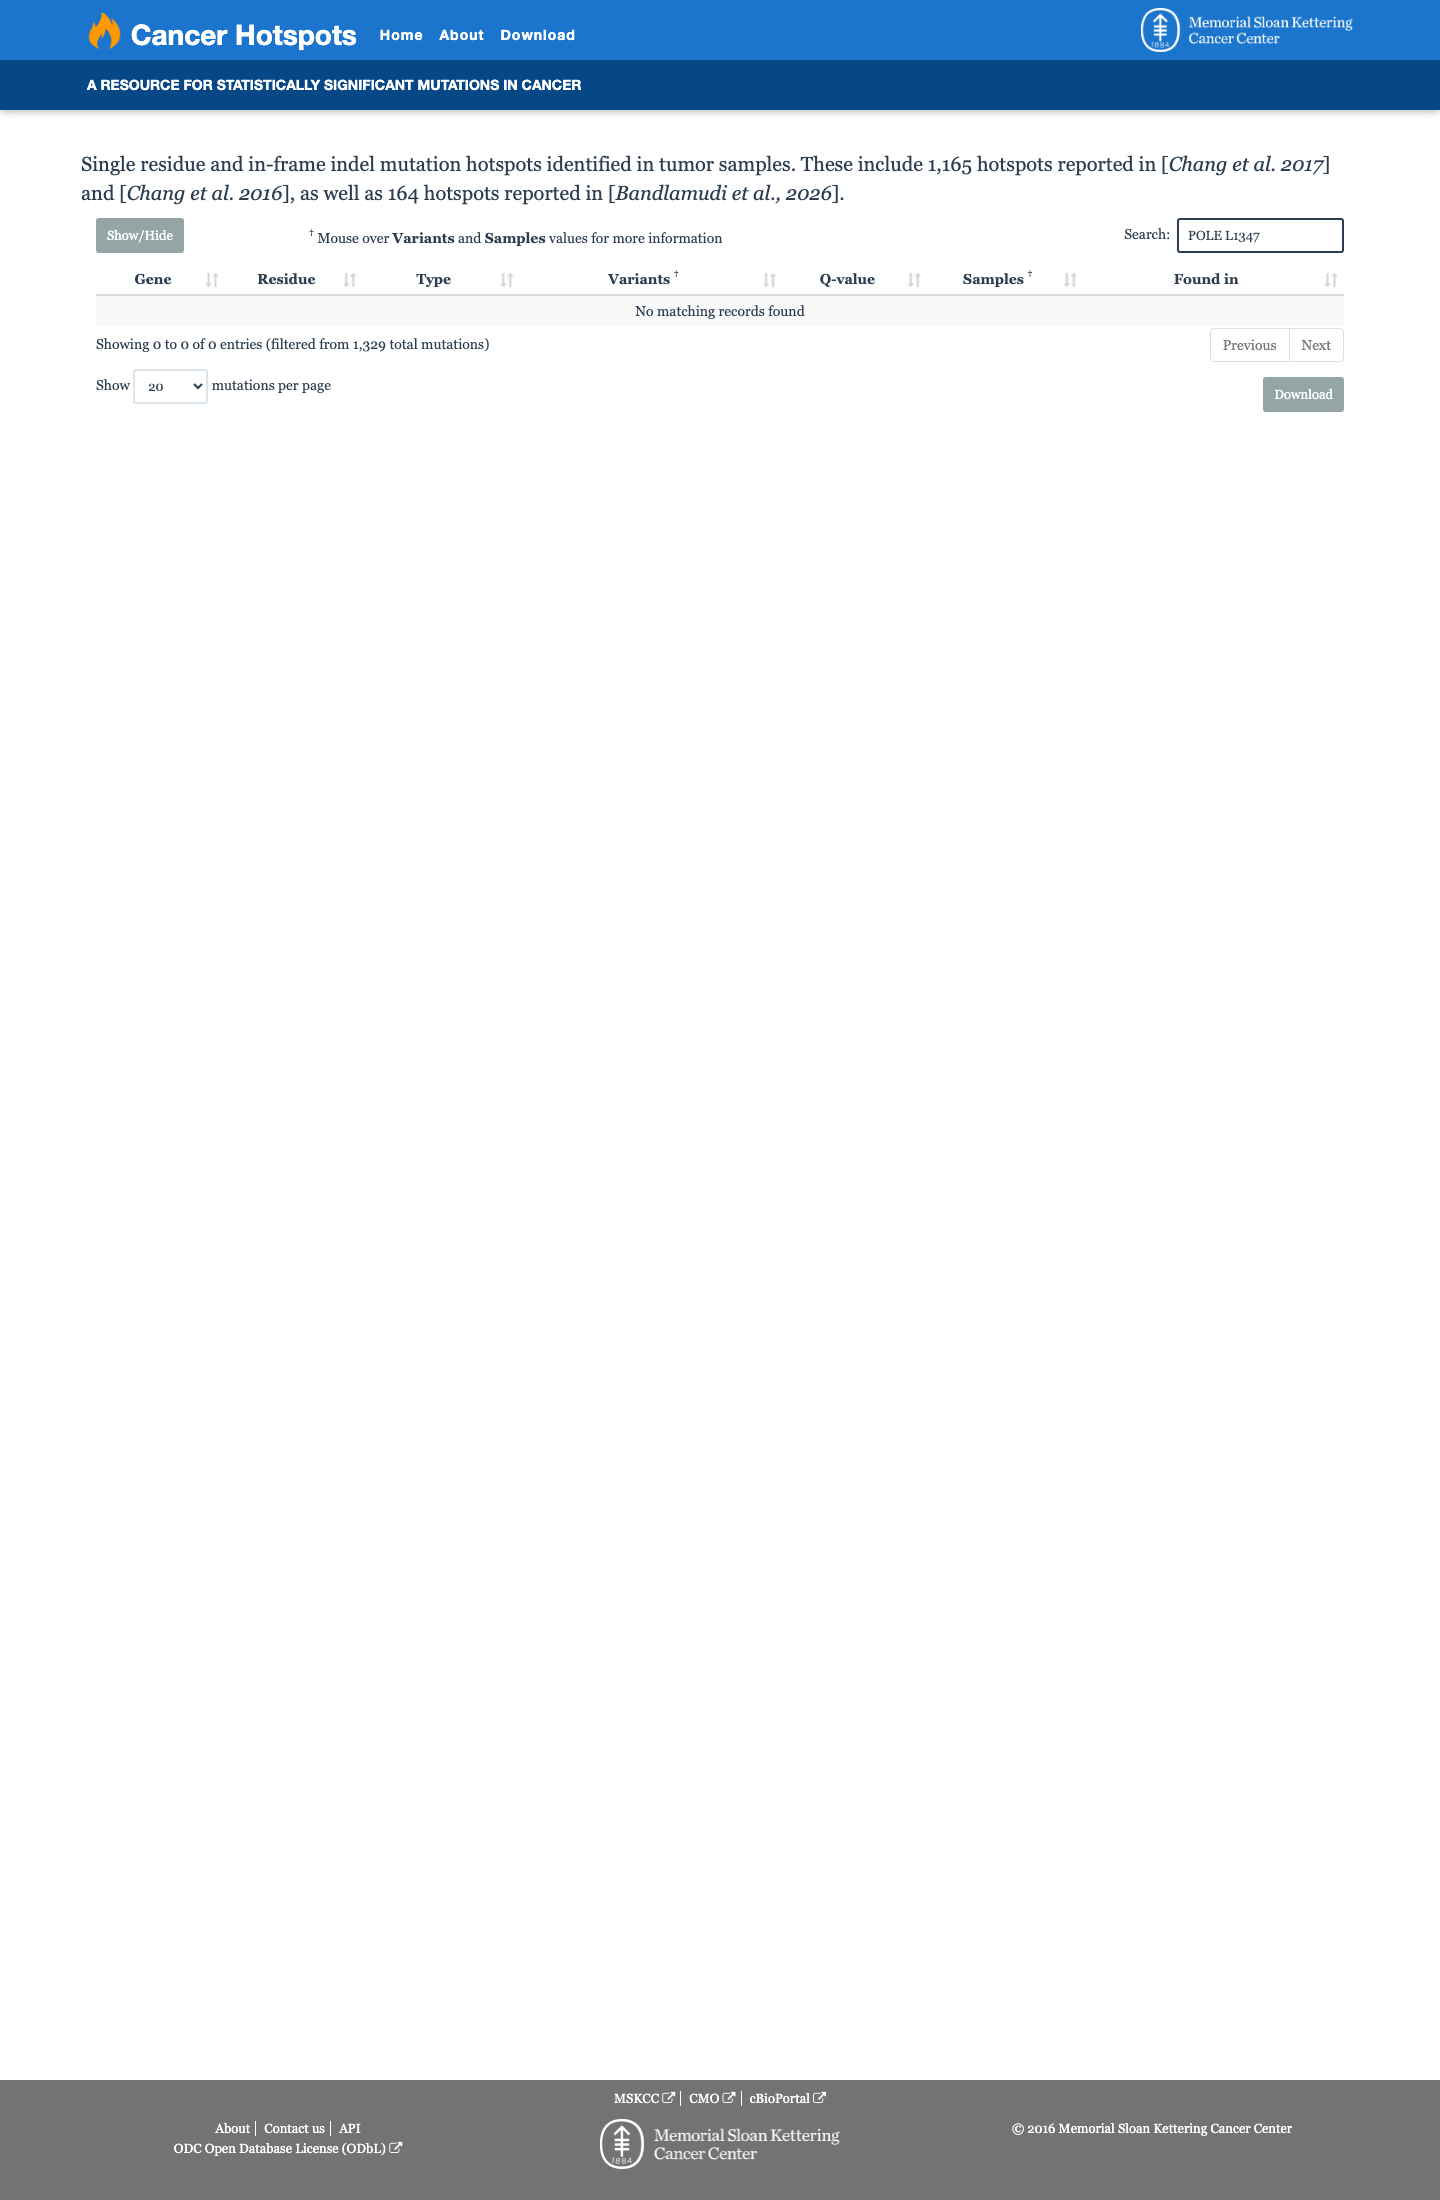
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
1papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 1 further PMID triaged but not cited — see Sources & references.
Rule & framework references · cited for criterion definitions, not variant evidence
25394175 ↗ A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 1 PMID not cited in assessment
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification criteria. CLINVAR