Back
NM_006231.4:c.1100T>G
p.Phe367Cys · POLE
León-Castillo et al. 2020 custom POLE framework · vleon-castillo-2020-custom-framework-v1
0%
complete
Final classification
VUS
PM2PM5PP3
POLE
c.1100T>G
p.Phe367Cys
missense · exon 11

POLE encodes the catalytic subunit of DNA polymerase epsilon, the enzyme that replicates the leading strand of DNA during cell division and participates in DNA repair. It contains a proofreading domain that corrects replication errors, keeping the accumulation of mutations in check. Germline mutations in POLE cause polyposis and predispose to colorectal cancer, and are also linked to a rare syndrome of facial dysmorphism, immunodeficiency, livedo, and short stature. Somatic mutations, particularly in the proofreading domain, occur in colorectal and endometrial cancers, where they drive an ultra-mutated tumor phenotype and are associated with better responses to immune checkpoint inhibitors.

This variant

This POLE missense variant affects the proofreading-domain protein encoded by a gene involved in leading-strand DNA replication and repair and associated with hereditary polyposis and colorectal-cancer predisposition.

Transcript
NM_006231.4
HGVS · transcript:coding
NM_006231.4:c.1100T>G
GRCh38
chr12:132675741 A>C
GRCh37
chr12:133252327 A>C
VUS: PM5 (moderate) plus PM2 and PP3 (supporting) do not meet the local POLE framework's Likely Pathogenic or Pathogenic combination rules.
Classification rationale
PM2PM5PP3 VUS
POLE c.1100T>G missense · exon 11

PM5 moderate: p.F367S is a pathogenic different missense change at the same residue. PM2 supporting: the variant is absent from gnomAD v2.1 and v4.1. PP3 supporting: the exact F367C row is REVEL likely disease causing with zero benign predictions.

PM2 + PM5 + PP3 → VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_006231.4 · variants mapped to exon structure
POLE NM_006231.4
Fetching transcript structure from UCSC…
Applied criteria · 3 applied · 21 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 3
Strength Supporting Moderate Strong Very strong
PM2 supporting Pathogenic
Met at Supporting: the variant is absent in gnomAD v2.1 and v4.1, meeting the PM2 threshold of <=0.0001.
gnomAD v2.1 all-comers: the variant was absent, corresponding to an observed allele frequency of 0.gnomAD v4.1 all-comers: the variant was absent, corresponding to an observed allele frequency of 0.The supplied generic PM2 calibration is <=0.0001 and specifies PM2 at Supporting strength under the ClinGen SVI recommendation (PMID:25741868).
PM5 moderate review Pathogenic
Met at moderate strength: p.F367S is a different missense change at residue 367 and was considered pathogenic in the TCGA-based analysis.
PATH-250-323.pdf, abstract: the six additional POLE mutations in the POLE-score >=4 group were considered pathogenic.PATH-250-323.pdf, results text: the six recurrent non-hotspot variants considered pathogenic include F367S.PATH-250-323-s002.xlsx, row p.F367(S^C): COSMIC 2, TCGA 2, total 4; this confirms a different missense substitution at the same residue 367.
PP3 supporting Pathogenic
Met at supporting: Supplementary Table S3 lists F367C as likely disease causing by REVEL with 0 benign in silico results.
León-Castillo et al. 2020 Supplementary Table S3, row F367C: SIFT=Deleterious; PANTHER=probably damaging; SNAP2=Effect; PolyPhen2=probably damaging; PROVEAN=Deleterious; REVEL=likely disease causing; Number of benign results=0.The exact variant was searched as c.1100T>G, p.Phe367Cys, and p.F367C; the governing in-silico table match is the F367C row.The PP3 rule requires an exact missense-table match, REVEL likely disease causing, and <=1 benign result; this row meets all three conditions.
Assessed · not applied · 7 not met · 14 not assessed
Pathogenic
PS1 Not assessed: no previously established pathogenic Phe367Cys variant caused by a different nucleotide change was documented in the reviewed sources.
PS2 Not assessed: no documented proband phenotype, family-history status, or confirmed maternity and paternity testing is available.
PS3 Not assessed: no validated functional assay result was identified; OncoKB's likely-loss-of-function label is not a documented assay outcome.
PS4 Not met: Supplementary Table S1 reports p.F367C in 1 COSMIC case, 0 TCGA cases, and 1 total, below the >=10 and dual-cohort recurrence requirements.
PM1 Not met: p.F367C is absent from the framework's exact PM1 lists, despite one COSMIC occurrence and zero TCGA occurrences in Supplementary Table S1.
PM3 Not assessed: no recessive inheritance or parental/offspring phase data show this variant in trans with a pathogenic variant.
PM6 Not assessed: no affected proband or parental-origin evidence supports an assumed de novo occurrence without confirmed maternity and paternity.
PP1 Not assessed: no affected family members or segregation results are documented for evaluating co-segregation with disease.
PP2 Not assessed: the reviewed sources provide no validated benign-missense-rate and common-missense-mechanism evidence for POLE.
PP4 Not assessed: no patient phenotype or family-history information is available to evaluate specificity for a single-gene disease.
PP5 Not met: ClinVar has no exact-variant expert-panel Pathogenic or Likely pathogenic classification for c.1100T>G.
Benign
BA1 Not met: the variant is absent from gnomAD v2.1 and v4.1, below the BA1 allele-frequency threshold of >=0.05.
BS1 Not met: gnomAD v2.1 and v4.1 show no observed allele, versus the generic BS1 threshold of >=0.01.
BS2 Not assessed: no healthy-adult carrier or homozygote observation is available for the BS2 zygosity and penetrance requirement.
BS3 Not assessed: no validated functional result showed absence of damage to POLE protein function or splicing; the available record instead reports likely loss of function.
BS4 Not assessed: no affected relatives and no family genotypes are documented to demonstrate lack of segregation.
BP1 Not assessed: the reviewed sources do not establish that POLE disease is caused primarily by truncating variants.
BP2 Not assessed: no phase data identify this variant in trans or cis with a pathogenic variant, and inheritance and penetrance are unestablished.
BP4 Not met: Supplementary Table S3 reports REVEL likely disease causing and 0 benign results, versus the BP4 requirement of Likely benign and >=4.
BP5 Not assessed: no alternate molecular basis for the patient's disease is documented in the available case evidence.
BP6 Not met: ClinVar has no exact-variant expert-panel Benign or Likely benign classification for c.1100T>G.
N/A · 4 PVS1 · PM4 · BP3 · BP7
Research & evidence
Population frequency
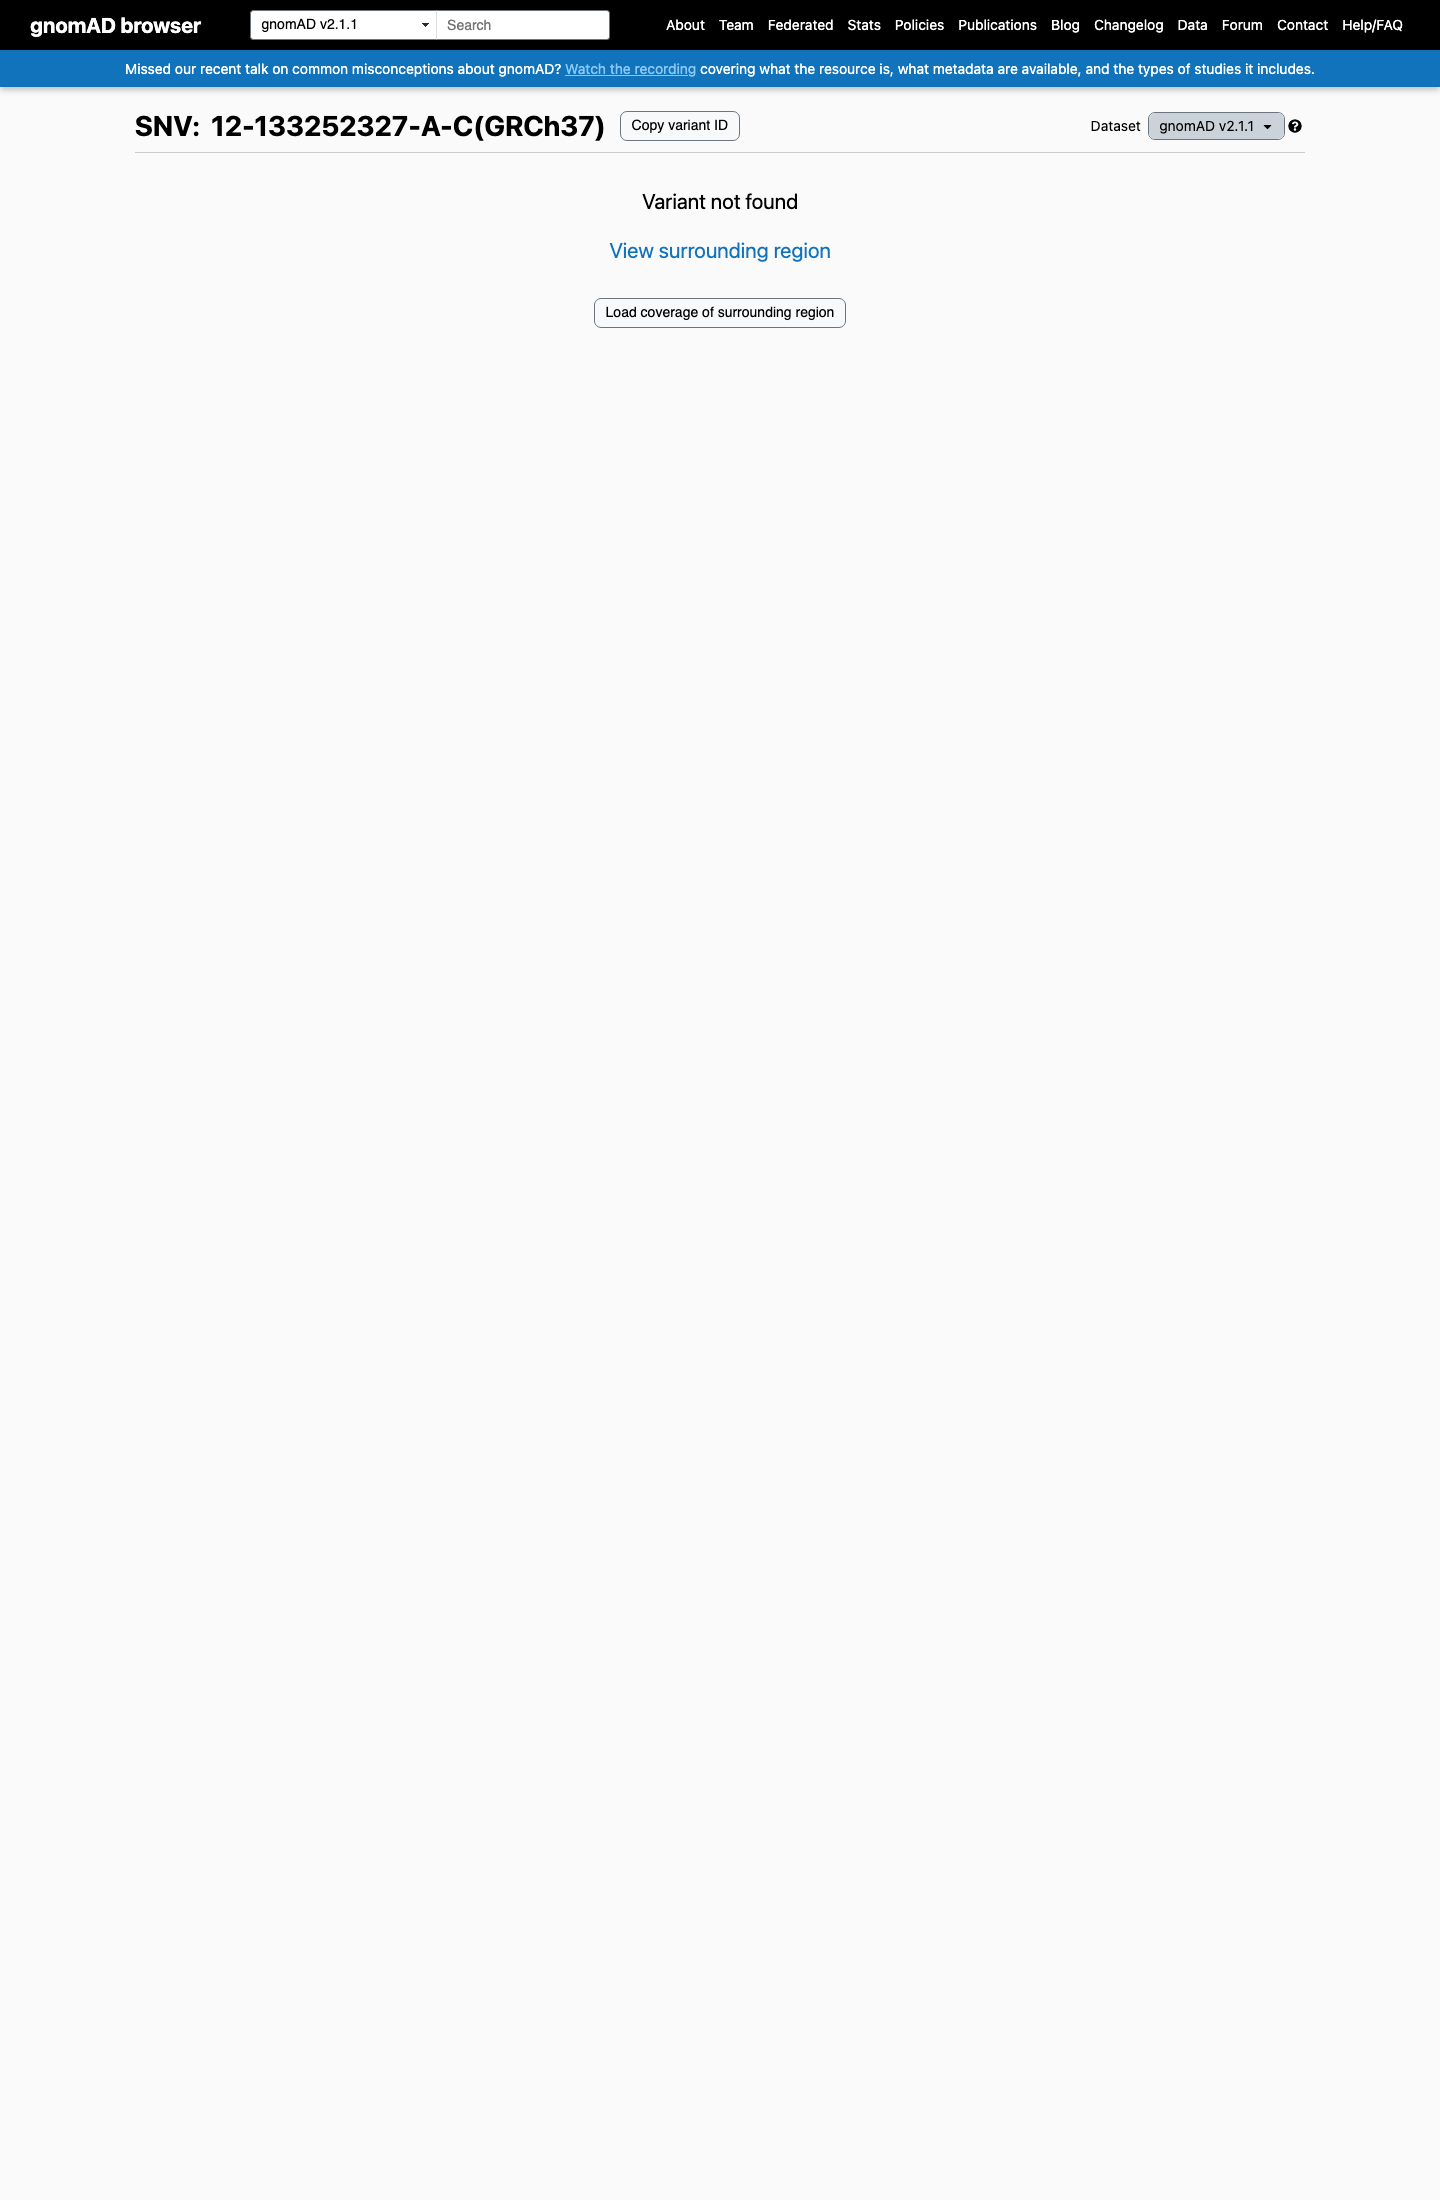
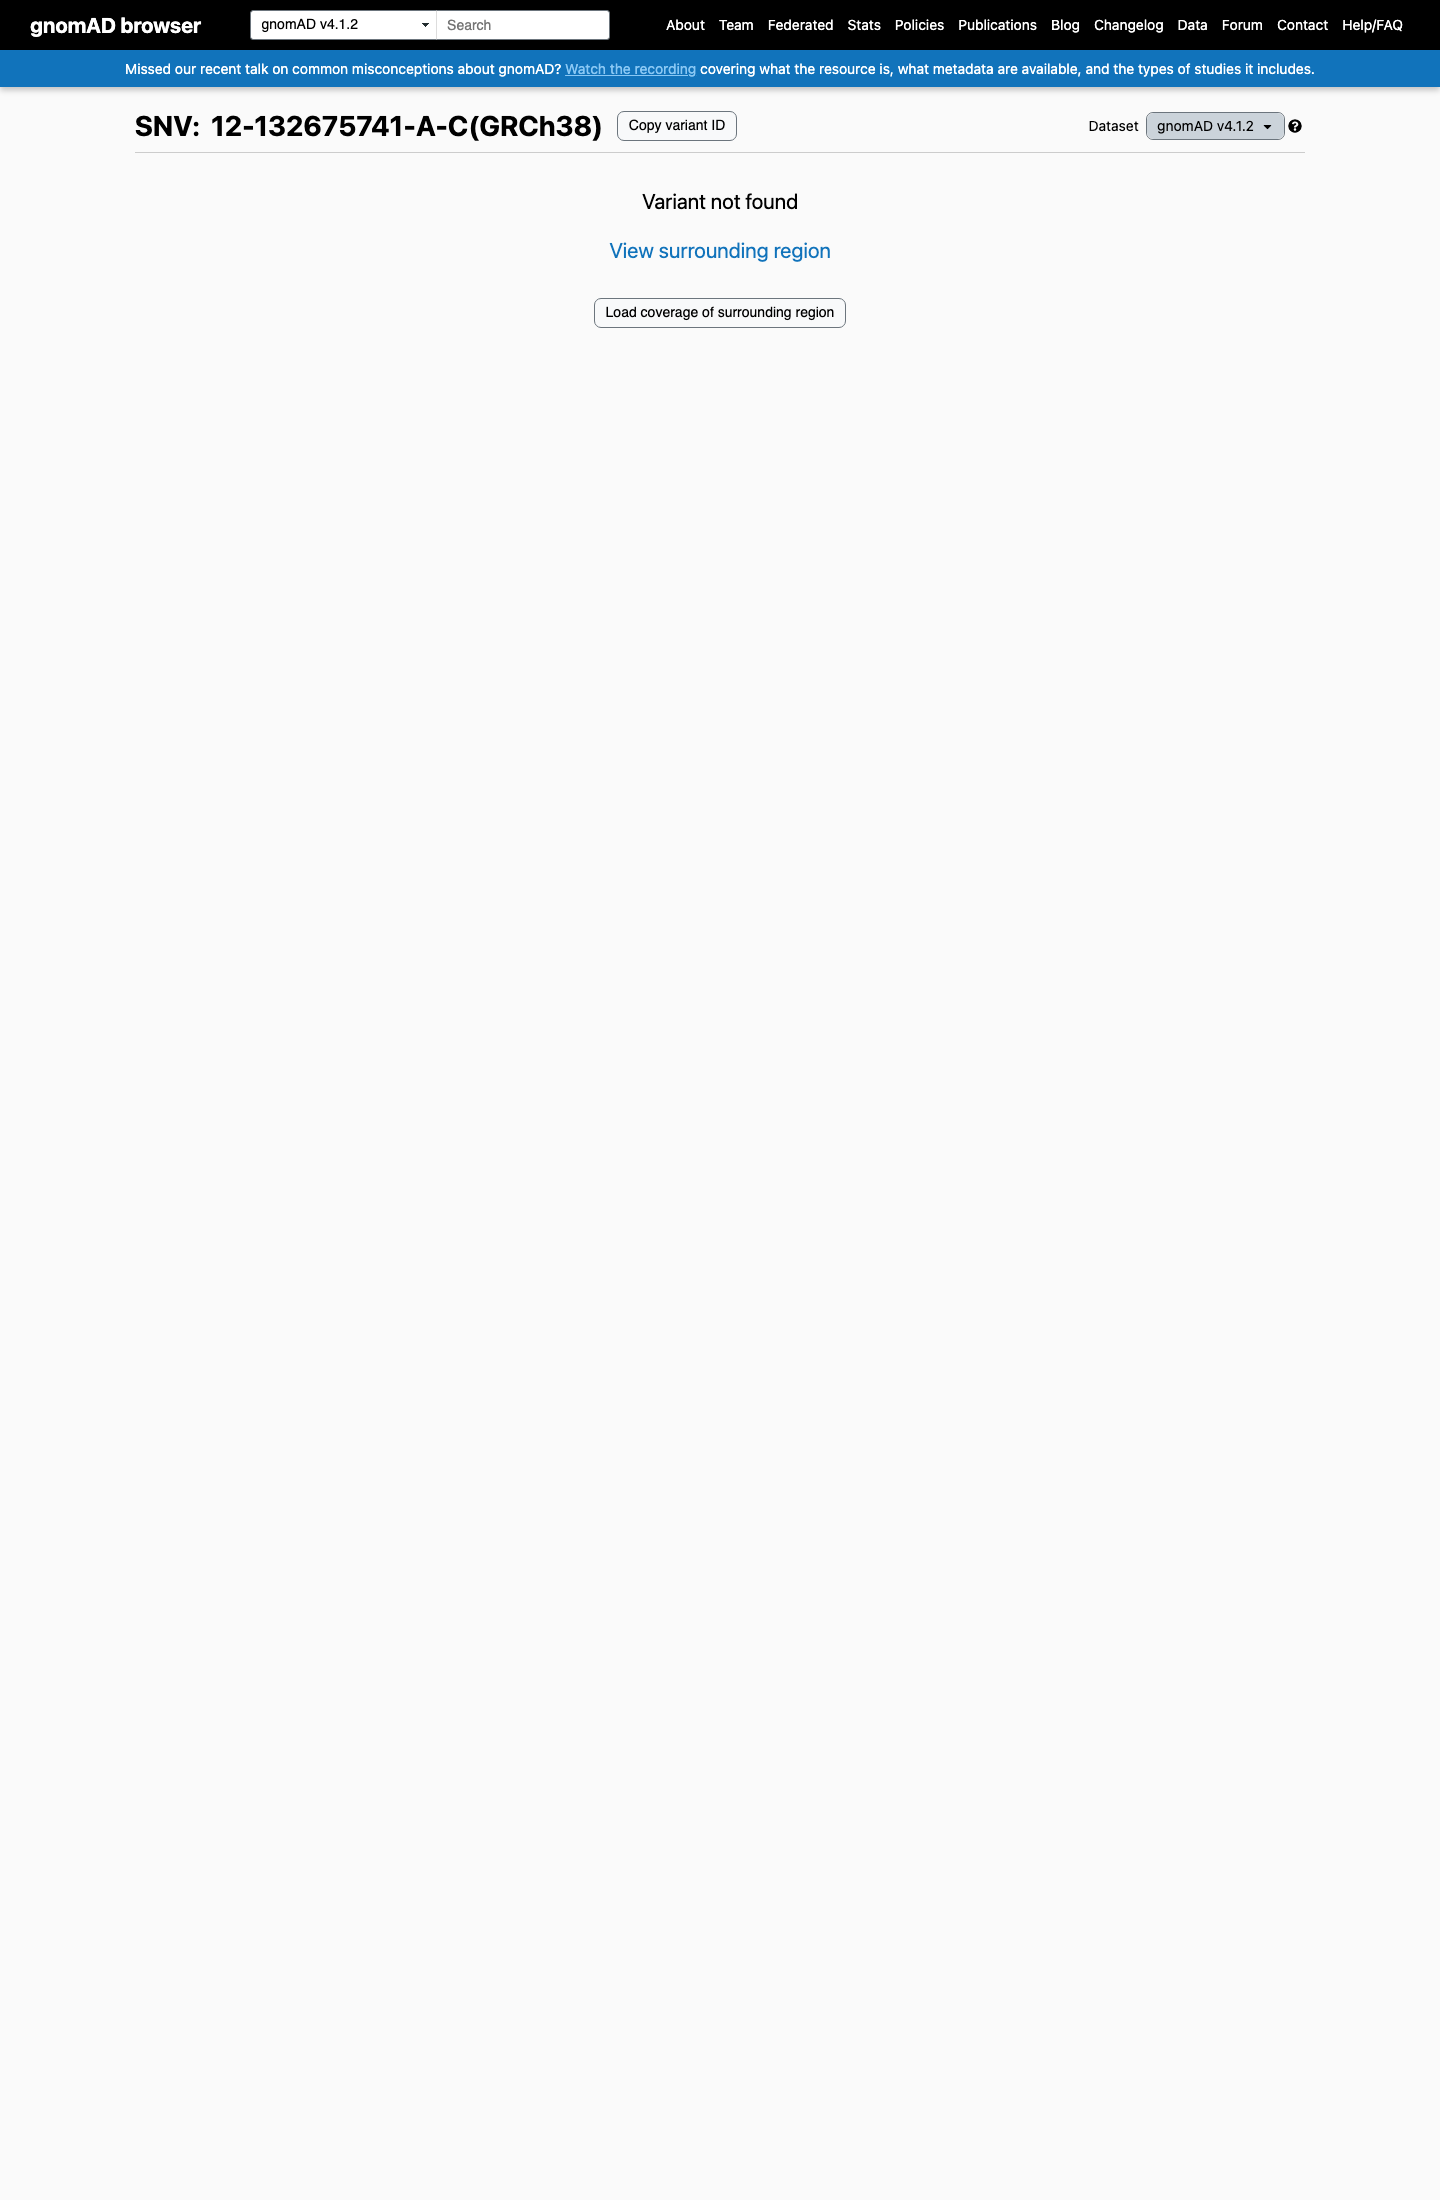
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
Absent from gnomAD v4.1.
v2.1
Absent from gnomAD v2.1.
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
gnomAD v2.1
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
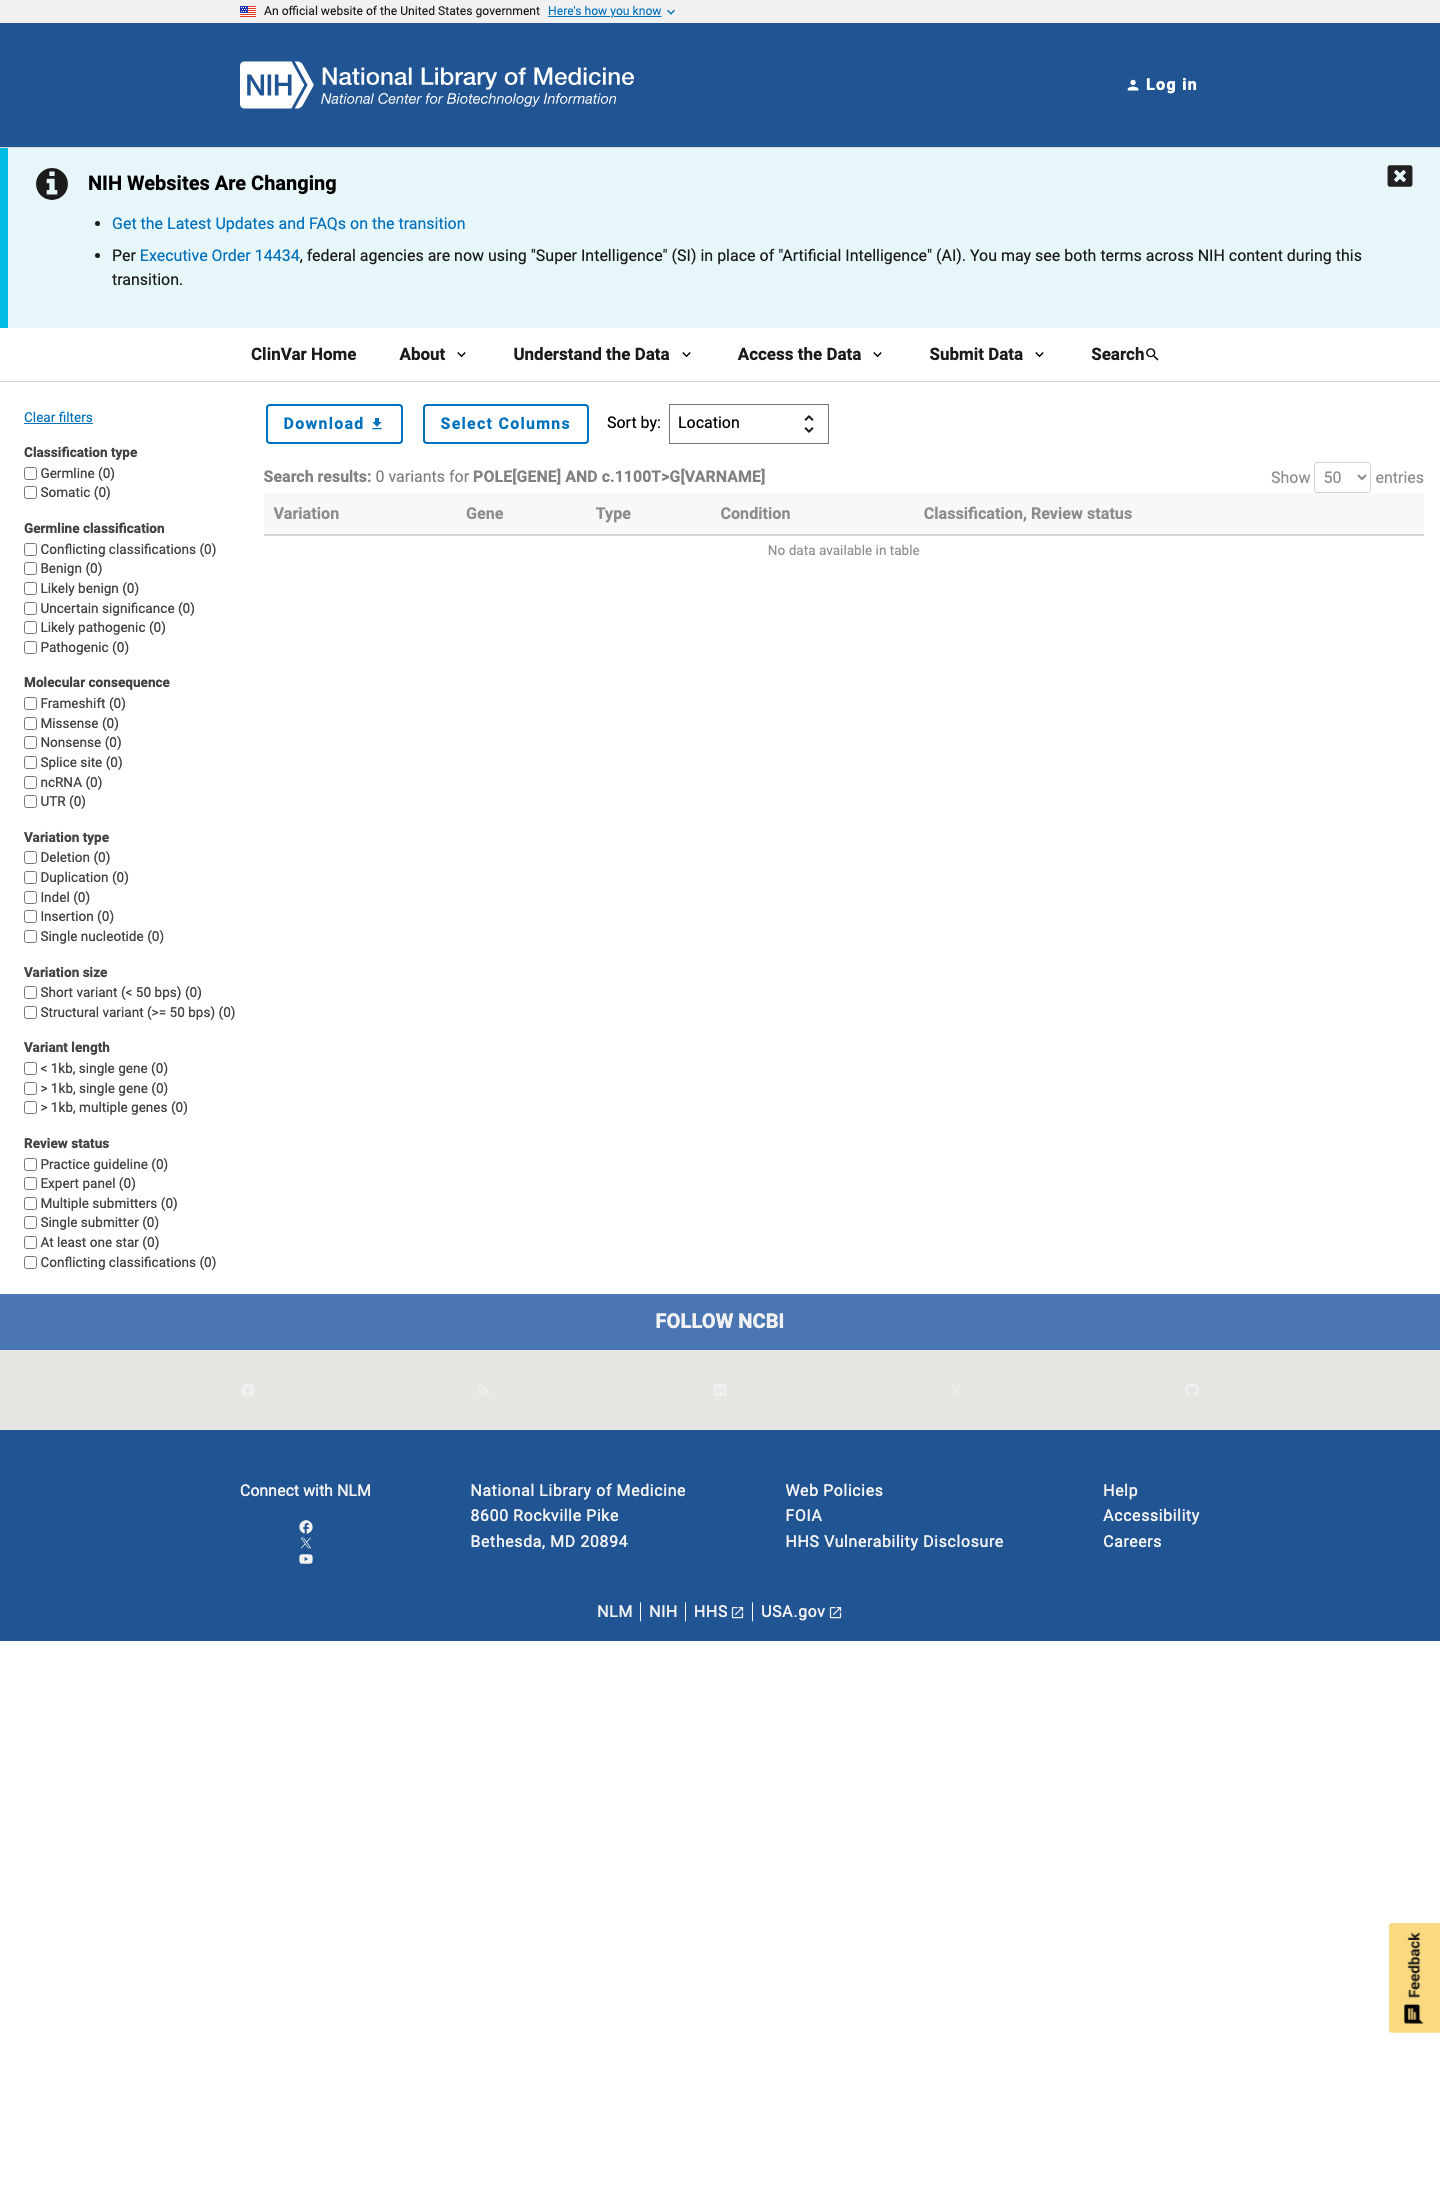
ClinVar screenshot
ClinVar
This variant is absent from ClinVar.
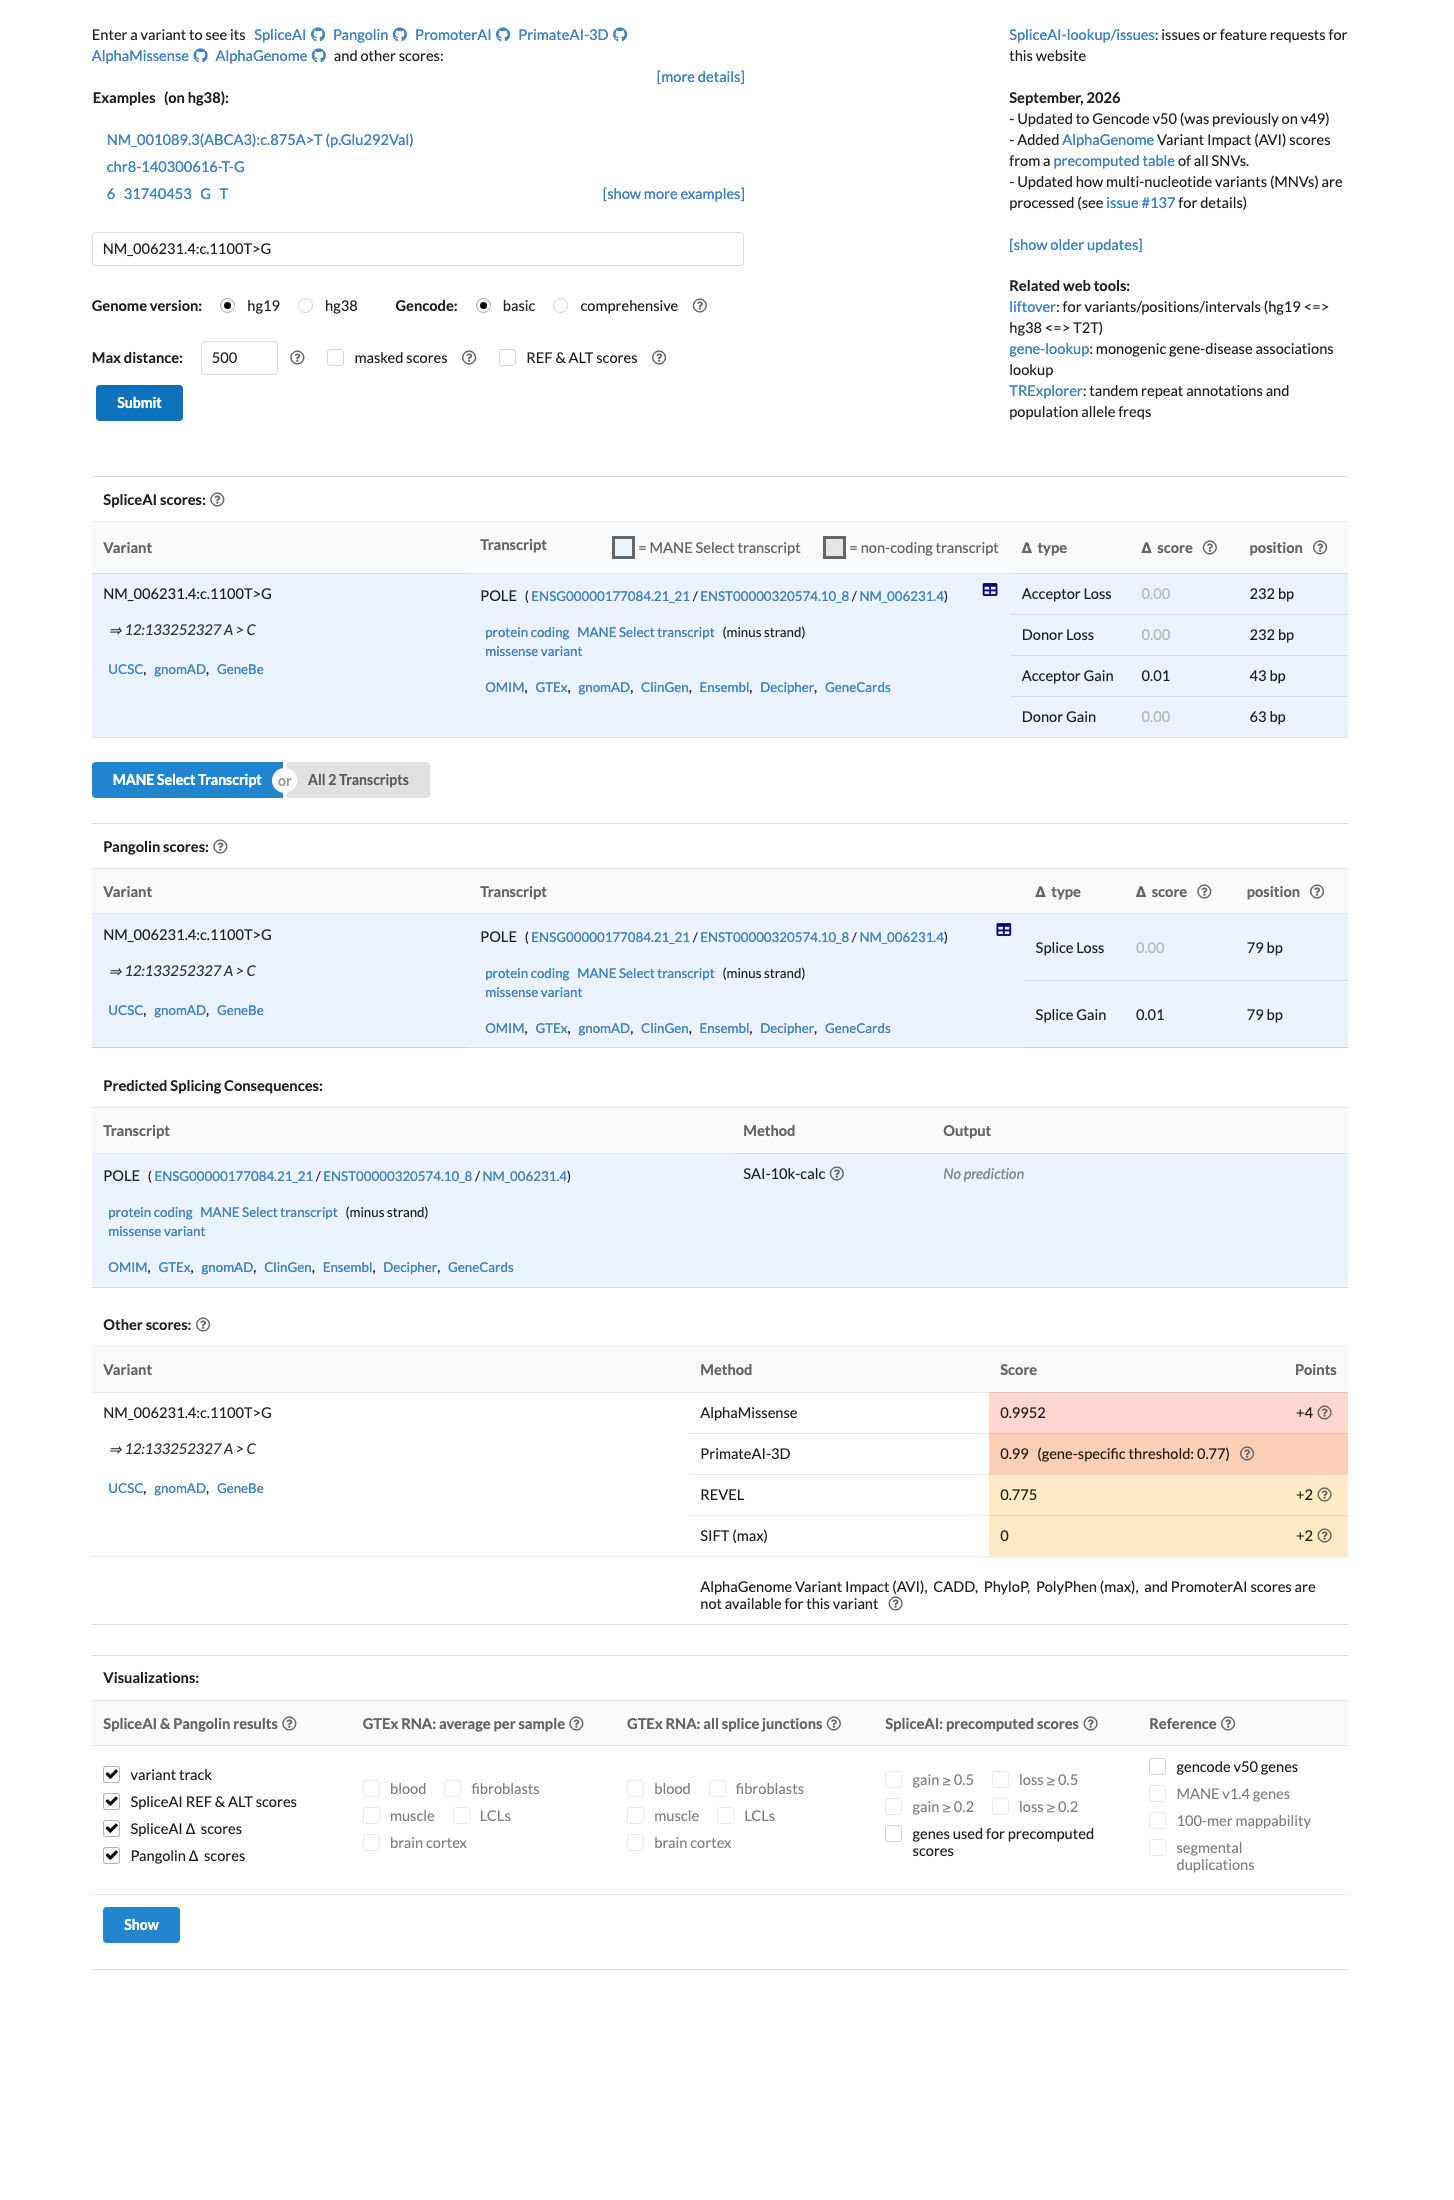
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.01). REVEL score = 0.775. BayesDel score = 0.197644.
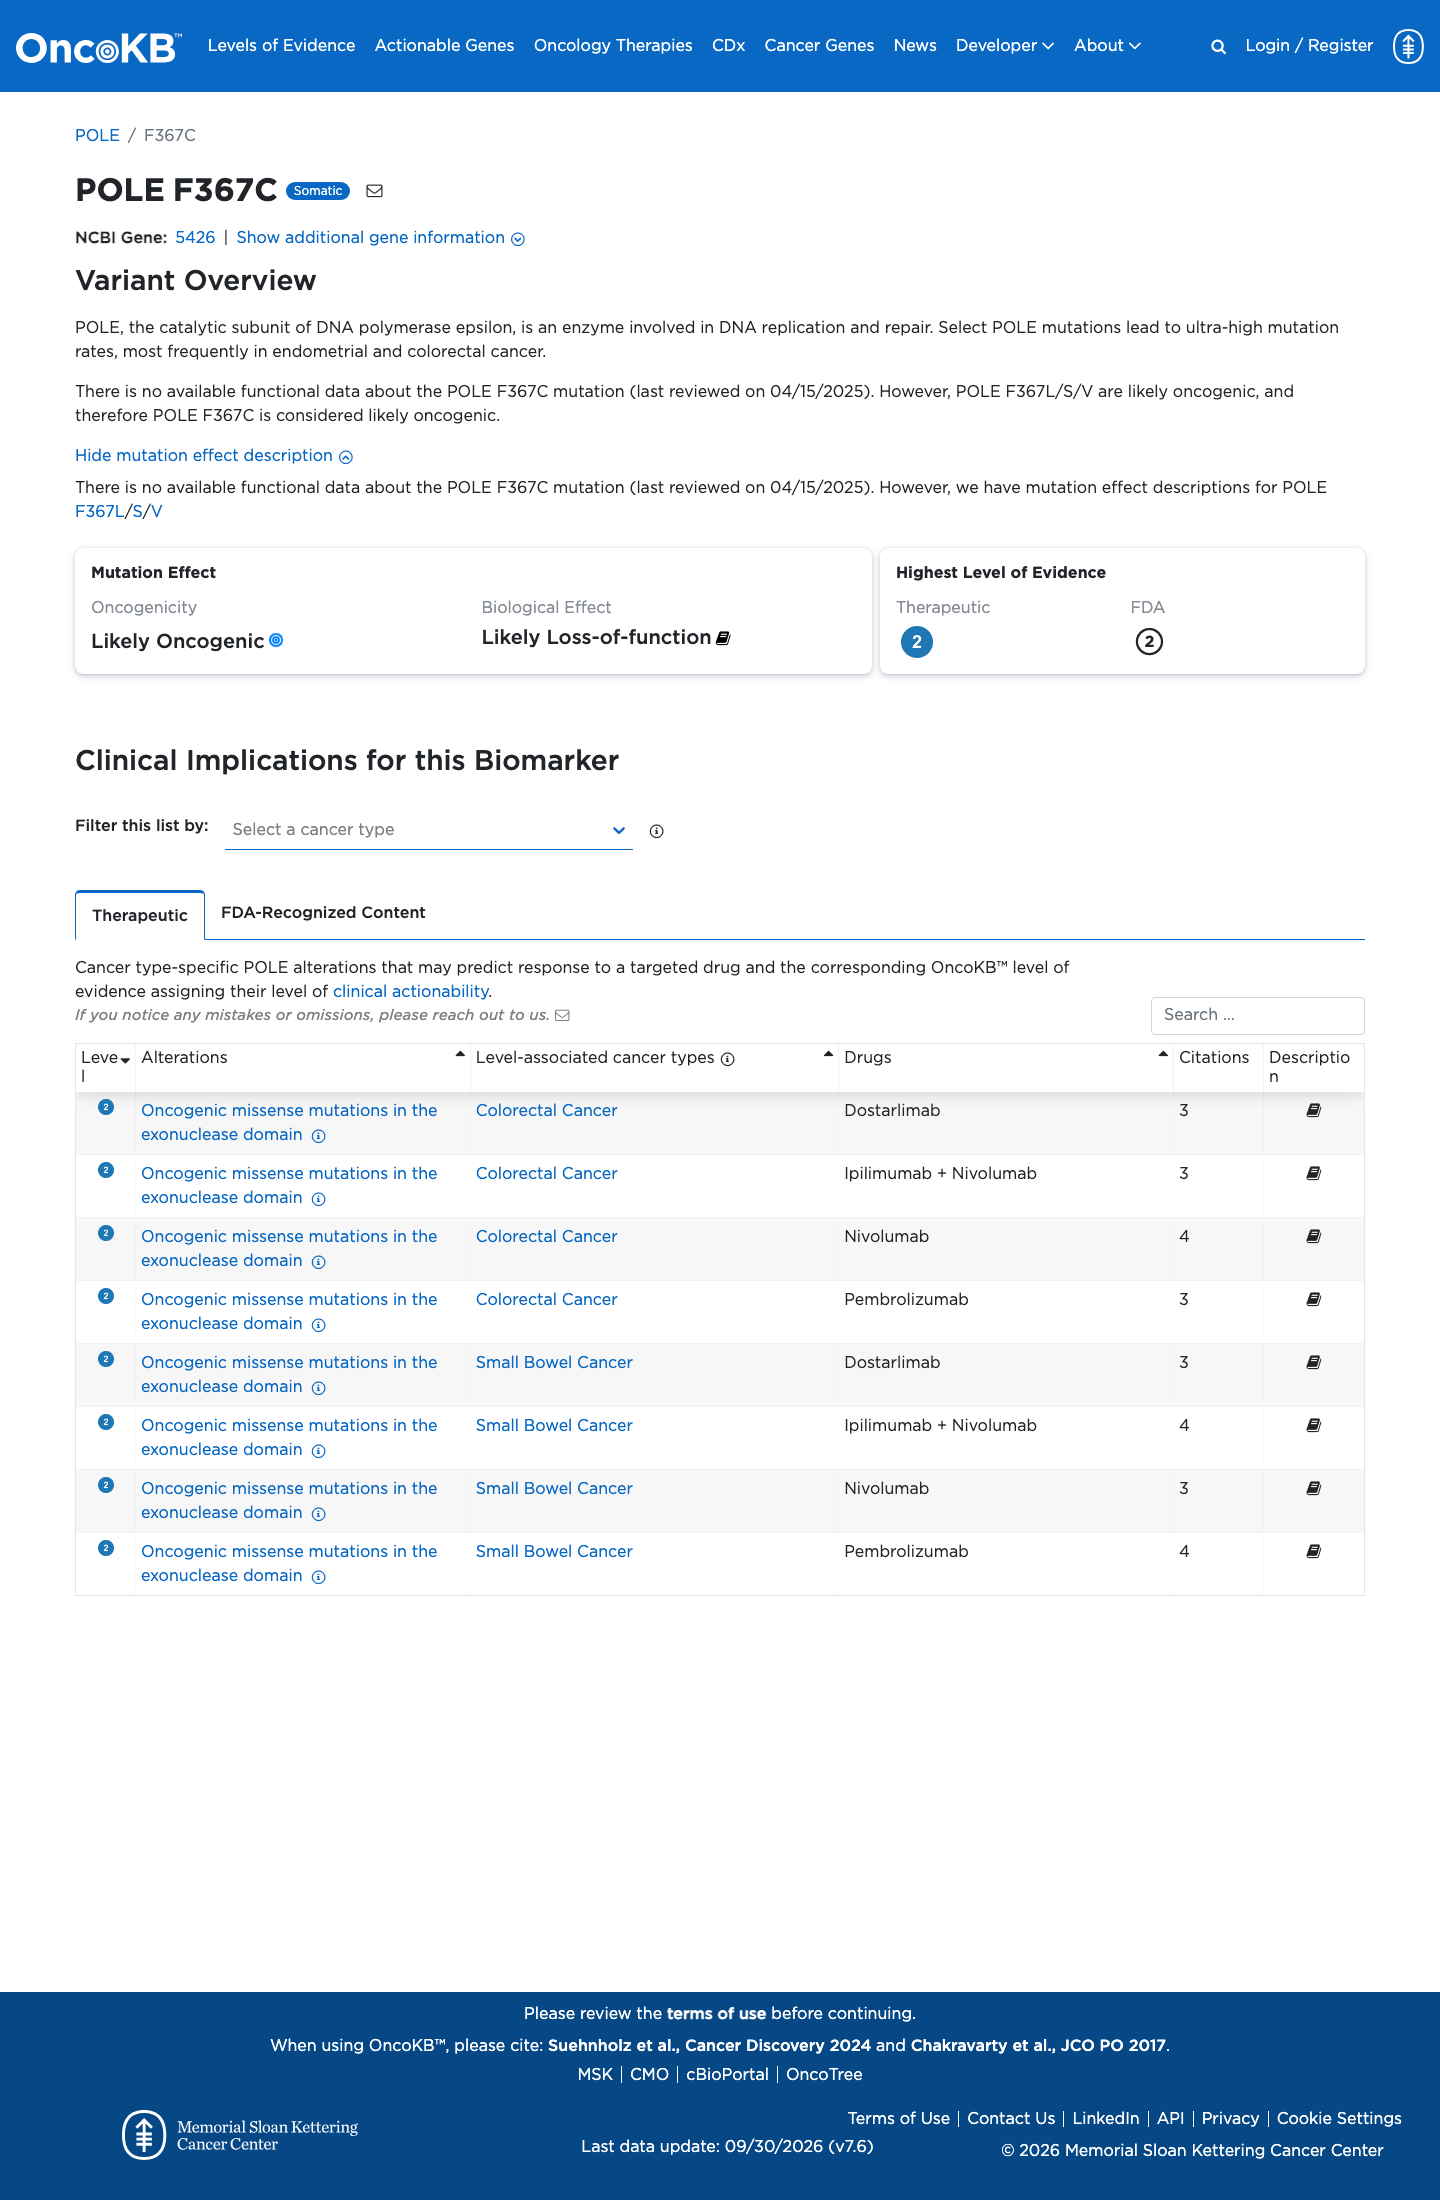
Functional / OncoKB screenshot
Functional Likely Oncogenic
OncoKB identified variant-specific curated literature and context relevant to functional review; biological-effect context: Likely Loss-of-function; curated oncogenicity label: Likely Oncogenic.
OncoKB ↗
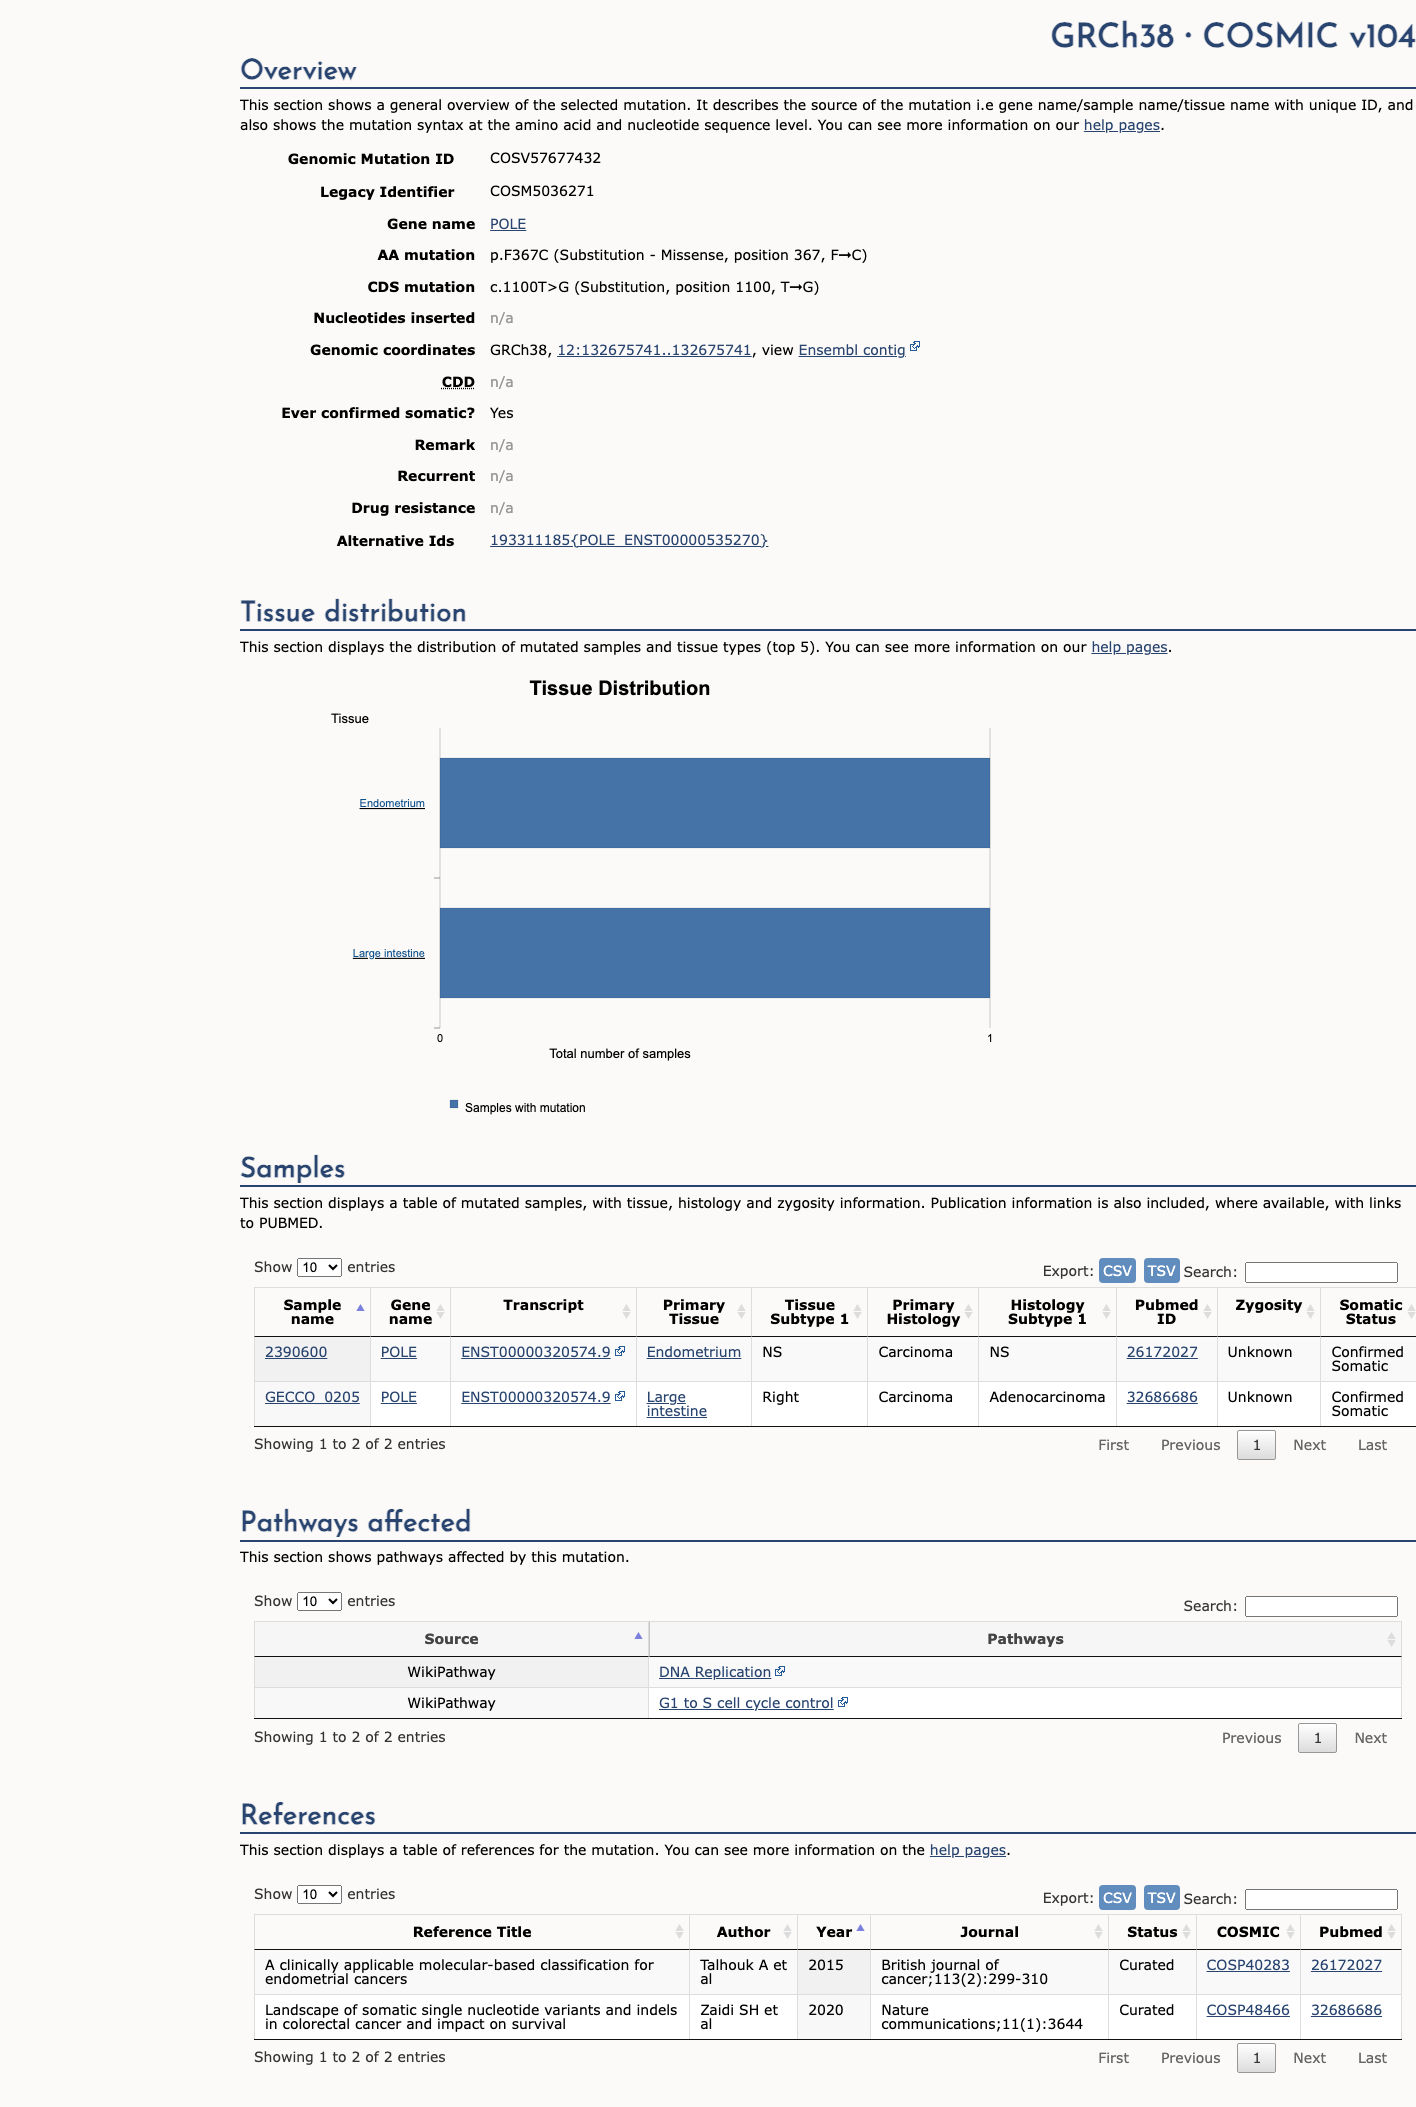
COSMIC screenshot
COSMIC
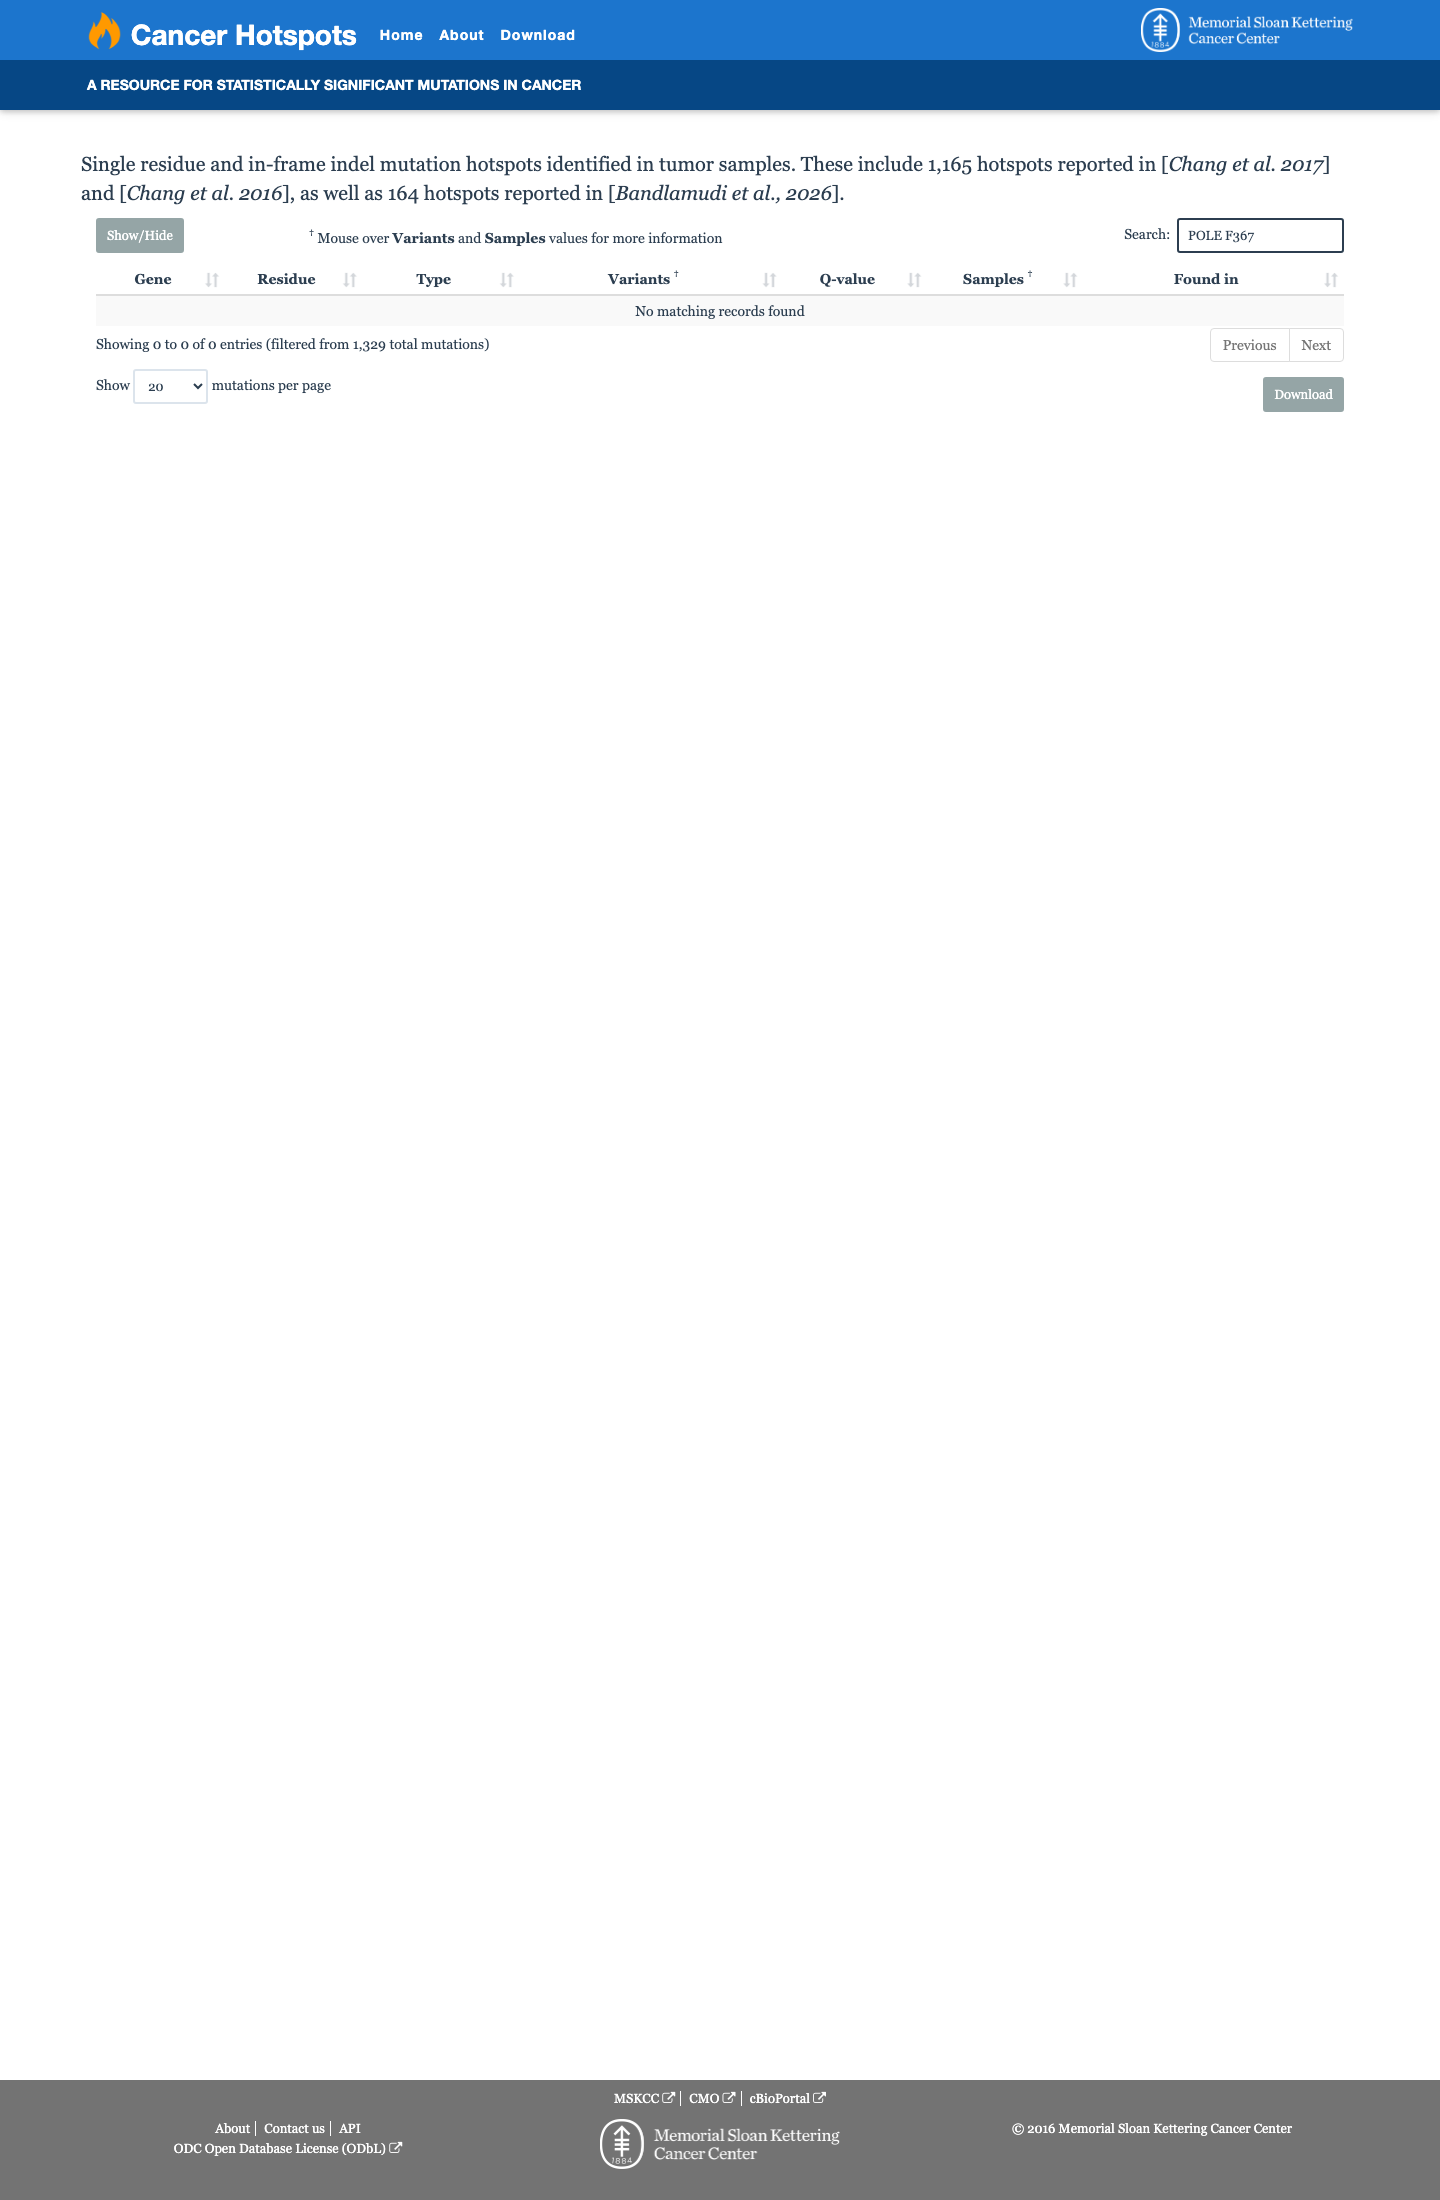
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has previously been reported in somatic cancers (COSMIC; COSV57677432, n = 2 times).
Hotspots
This variant does not lie in a statistically significant hotspot.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots