Back
NM_006231.4:c.2792T>C
p.Phe931Ser · POLE
León-Castillo et al. 2020 custom POLE framework · vleon-castillo-2020-custom-framework-v1
0%
complete
Final classification
VUS
PM2PP3
POLE
c.2792T>C
p.Phe931Ser
missense · exon 24

POLE encodes the catalytic subunit of DNA polymerase epsilon, the enzyme that replicates the leading strand of DNA during cell division and participates in DNA repair. It contains a proofreading domain that corrects replication errors, keeping the accumulation of mutations in check. Germline mutations in POLE cause polyposis and predispose to colorectal cancer, and are also linked to a rare syndrome of facial dysmorphism, immunodeficiency, livedo, and short stature. Somatic mutations, particularly in the proofreading domain, occur in colorectal and endometrial cancers, where they drive an ultra-mutated tumor phenotype and are associated with better responses to immune checkpoint inhibitors.

This variant

This POLE missense change alters a residue in the polymerase region of the protein, well outside the exonuclease proofreading domain whose germline defects account for POLE-associated polyposis and colorectal cancer predisposition.

Transcript
NM_006231.4
HGVS · transcript:coding
NM_006231.4:c.2792T>C
GRCh38
chr12:132661599 A>G
GRCh37
chr12:133238185 A>G
VUS: PM2 (supporting) and PP3 (supporting) are the only met criteria, and neither the POLE framework's combination rules nor generic ACMG/AMP reach a classification threshold.
Classification rationale
PM2PP3 VUS
POLE c.2792T>C missense · exon 24

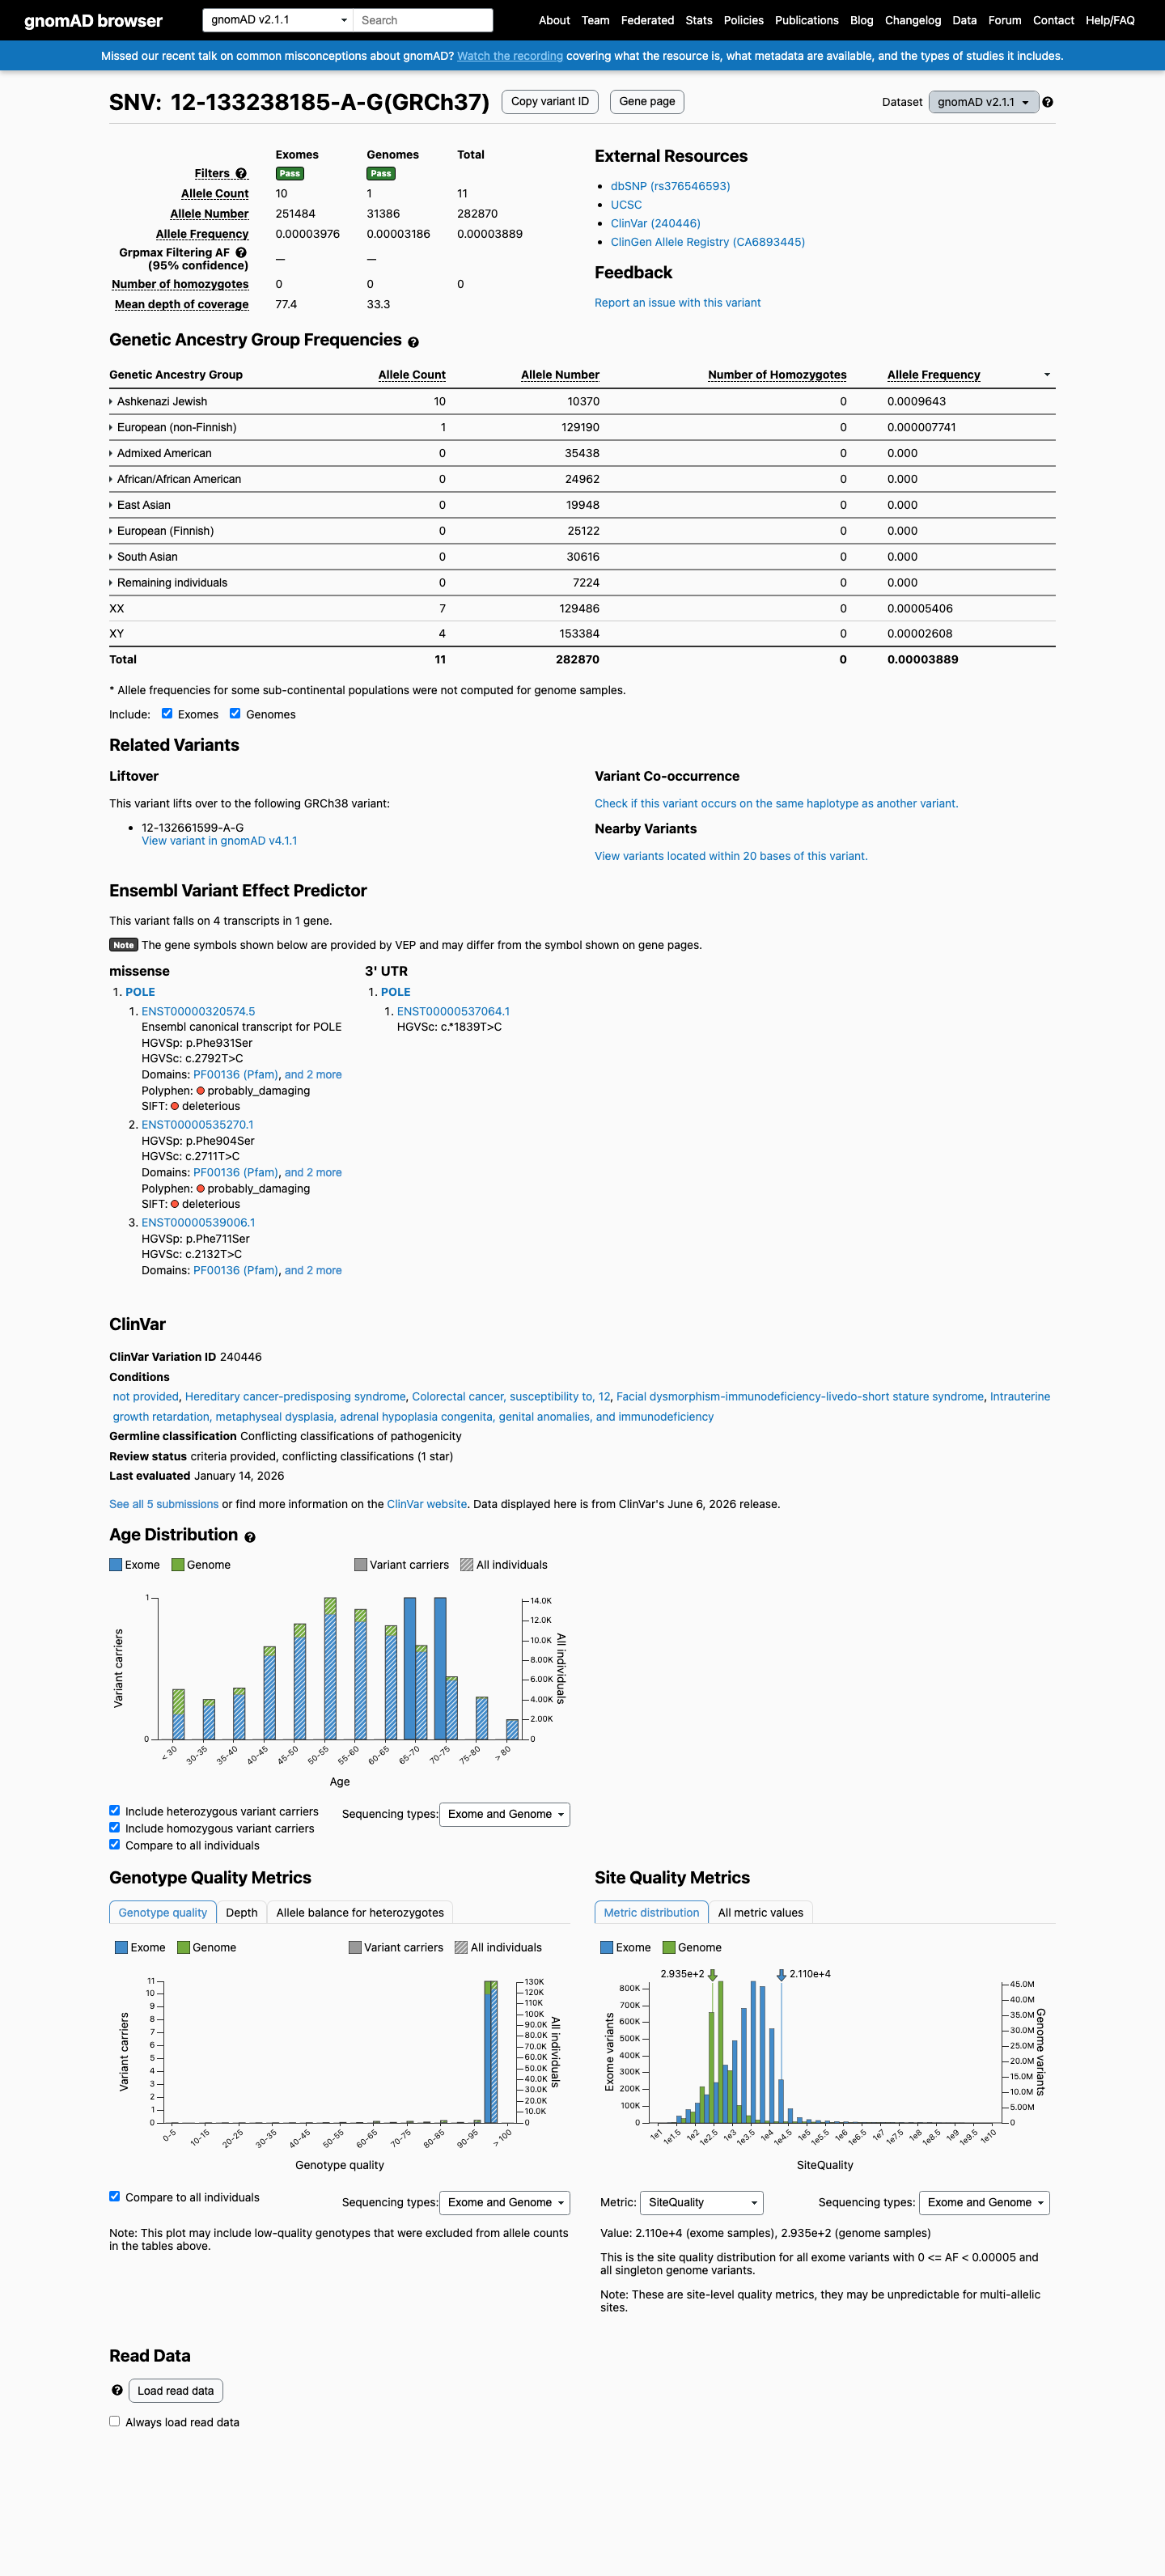
PM2 supporting: gnomAD v4.1 all-comers allele frequency is 1.67e-05, below the 0.0001 threshold, with no homozygotes. PP3 supporting: REVEL 0.749 places this missense change in the PP3 supporting band under the generic in-silico fallback.

PM2 + PP3 VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_006231.4 · variants mapped to exon structure
POLE NM_006231.4
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 22 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
PM2 supporting review Pathogenic
Met, supporting: gnomAD v4.1 all-comers AF 1.67e-05 is below the 0.0001 PM2 threshold, though Ashkenazi Jewish AF reaches 0.000777.
No ClinGen CSPEC/VCEP for POLE was available (cspec not found); the governing specification is the local Leon-Castillo et al. 2020 custom POLE framework (custom_internal, official_clingen false, precedence rank 2), which customizes only PM1, PS4, PP3 and BP4. It neither specifies a non-cancer or exome-only population source nor a frequency cutoff for PM2, so the default all-comers gnomAD v2.1 and gnomAD v4.1 entries govern, with generic ACMG/AMP thresholds.Generic PM2 threshold used: observed allele frequency <=0.0001, applied at supporting strength only (ClinGen SVI recommendation to downgrade PM2 to supporting, based on Richards et al. 2015, PMID:25741868).gnomAD v2.1 all-comers: total AF 3.8887121292466504e-05 (11/282,870 alleles; exome 10/251,484, genome 1/31,386), 0 homozygotes. This is below the 0.0001 threshold.
PP3 supporting Pathogenic
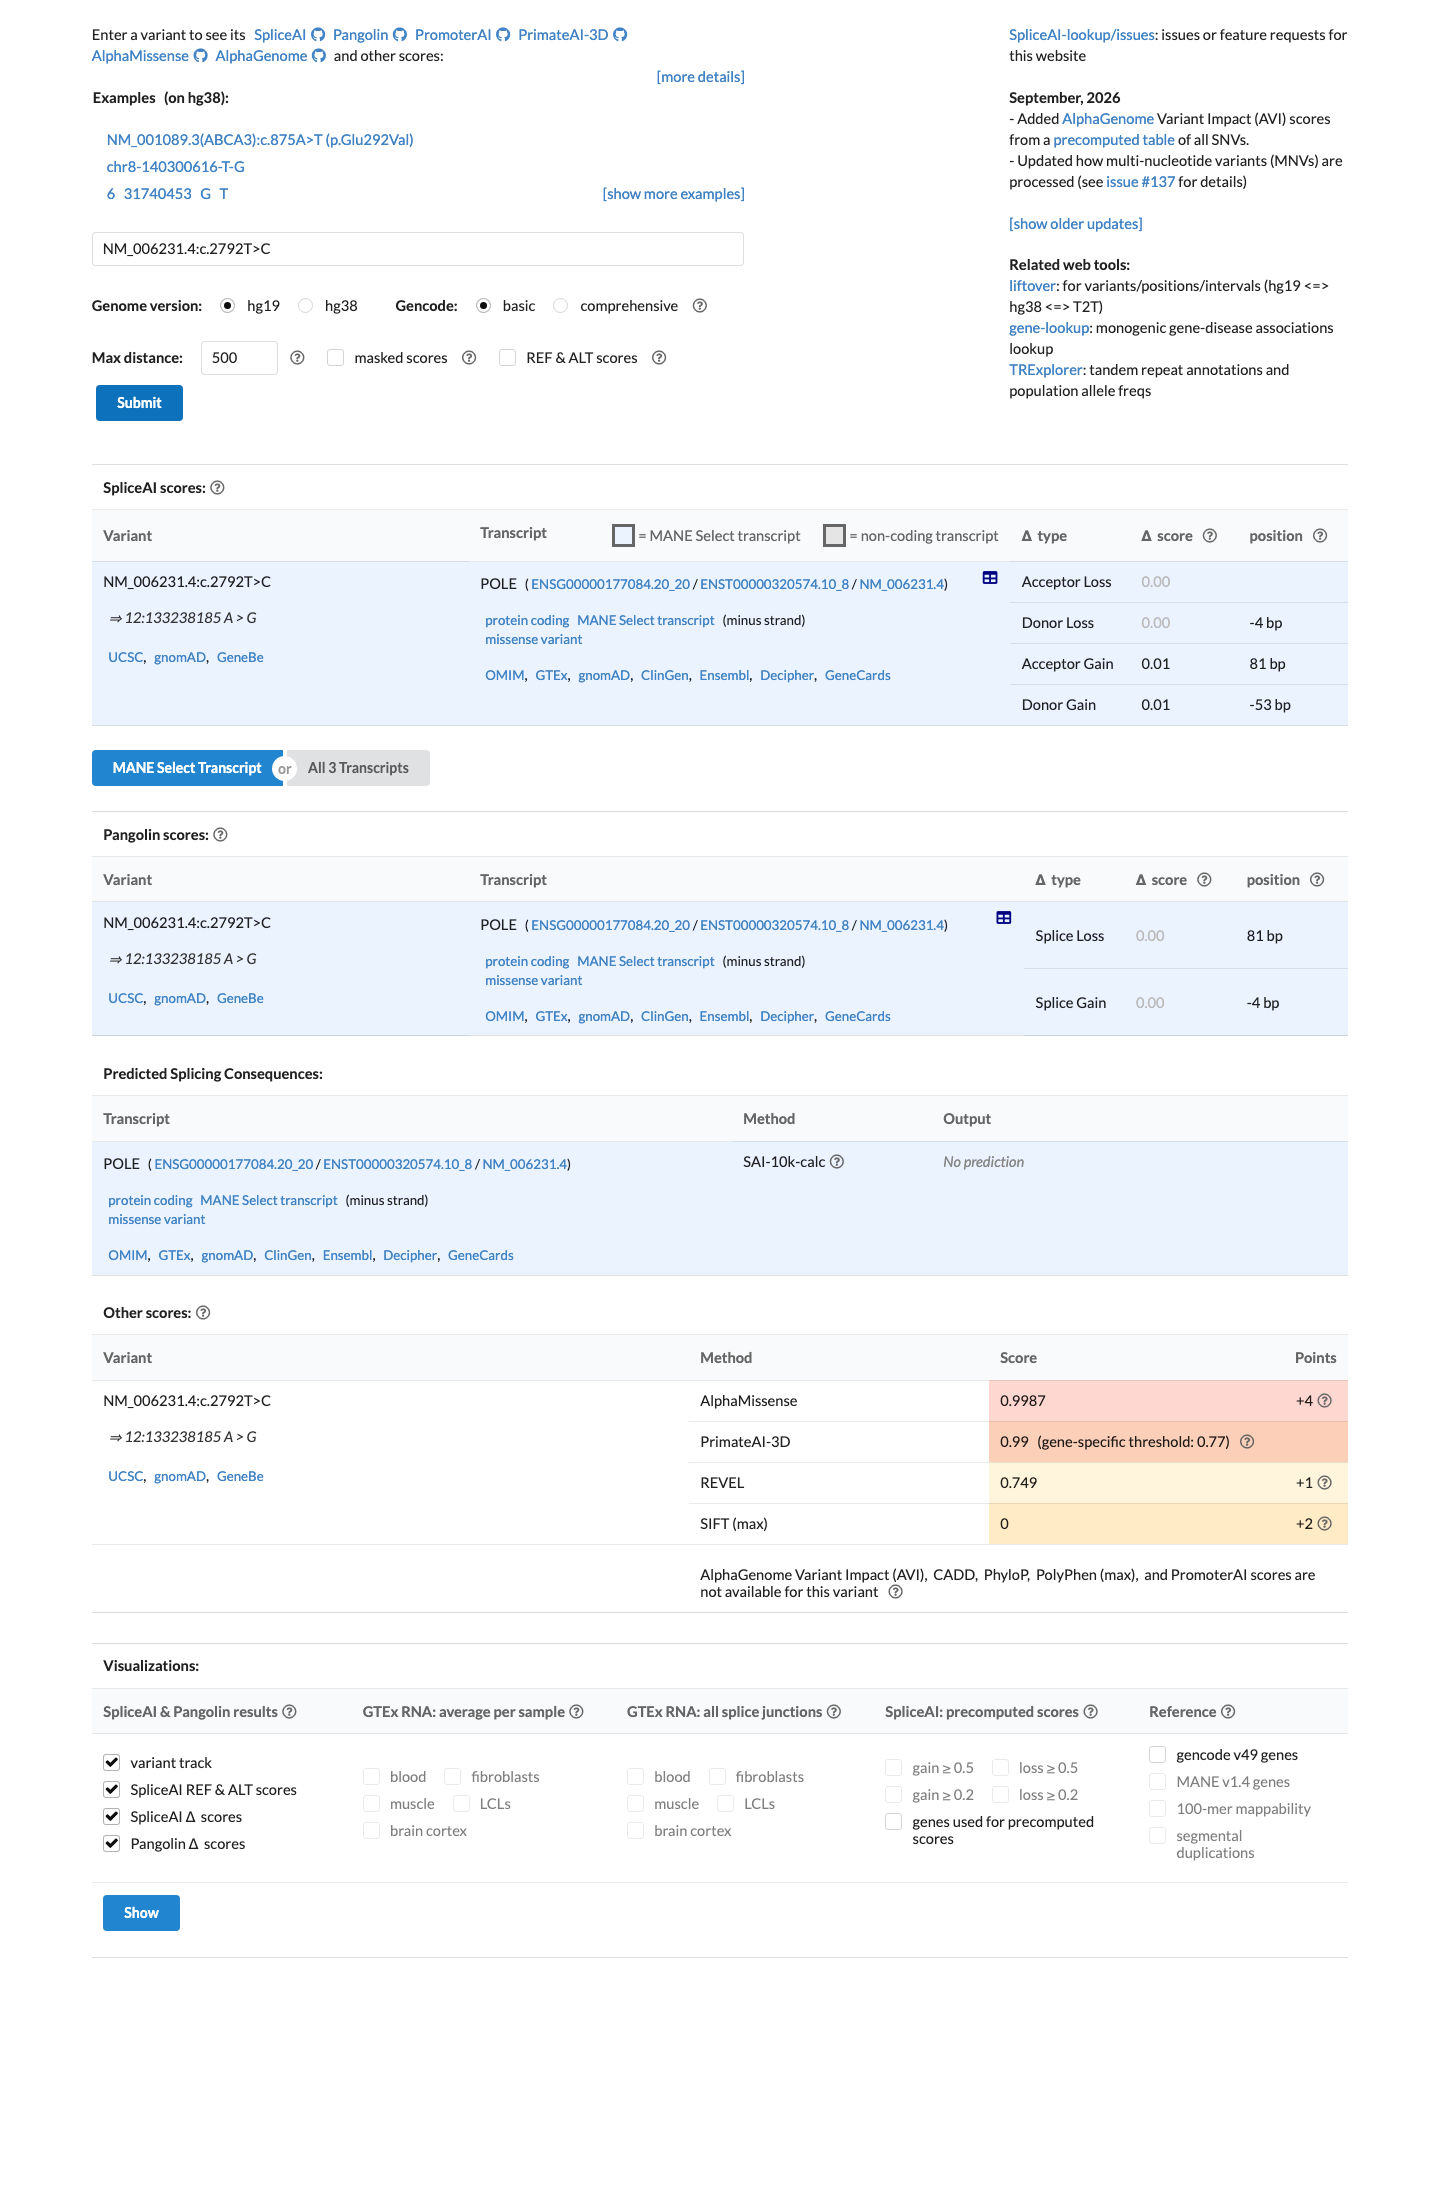
Met at supporting: REVEL 0.749 sits in the PP3-supporting band (>=0.644) and below the 0.773 moderate cutoff.
PP3 scope check: c.2792T>C is a missense substitution p.(Phe931Ser), so the criterion is in scope; PP3 was not evaluated as a splice-impact call (SpliceAI max delta 0.008, nearest acceptor 81 bp and nearest donor 53 bp away, i.e. not a splice-region variant).Local REVEL v1.3 lookup returned revel_score = 0.749 at chr12:132661599 A>G (hg38; chr12:133238185 A>G hg37), found = true.Applied the published ClinGen SVI REVEL calibration verbatim: PP3 supporting >= 0.644, PP3 moderate >= 0.773, PP3 strong >= 0.932 (Pejaver et al. 2022, Am J Hum Genet, PMID:36413997). 0.749 >= 0.644 but < 0.773, giving PP3 at supporting strength.
Assessed · not applied · 11 not met · 11 not assessed
Pathogenic
PS1 Not met: no established pathogenic allele produces p.Phe931Ser, and the only codon-931 ClinVar record (240446) is uncertain significance, not pathogenic.
PS2 Not assessed: no proband or parental testing exists, so the confirmed de novo occurrence PS2 requires cannot be evaluated.
PS3 Not assessed: no functional or proofreading assay of POLE p.Phe931Ser was identified, leaving PS3's damaging-effect requirement unevaluable.
PS4 Not met: F931S is absent from Supplementary Table S1 (combined endometrial-cancer count 0, not >=10), so the POLE custom PS4 recurrence rule cannot fire.
PM1 Not met: p.F931S lies outside the POLE exonuclease domain (residues 268-471), where the governing framework restricts PM1 to 14 named hotspot/domain substitutions.
PM3 Not assessed: no trans-partner or phase data exists for heterozygous POLE c.2792T>C, so the recessive in-trans requirement cannot be evaluated.
PM5 Not met: zero codon-931 pathogenic or likely pathogenic comparators exist in ClinVar or the VCEP recurrence/in-silico tables.
PM6 Not assessed: no proband or parental information is reported, so an assumed de novo occurrence cannot be recorded.
PP1 Not assessed: no pedigree, affected relatives, or countable meioses are reported, so co-segregation cannot be scored.
PP2 Not assessed: no POLE-specific PP2 rule and no gene-level missense-constraint data were available to test the criterion.
PP4 Not assessed: no proband phenotype or family history was captured for this case, so phenotype specificity for POLE cannot be evaluated.
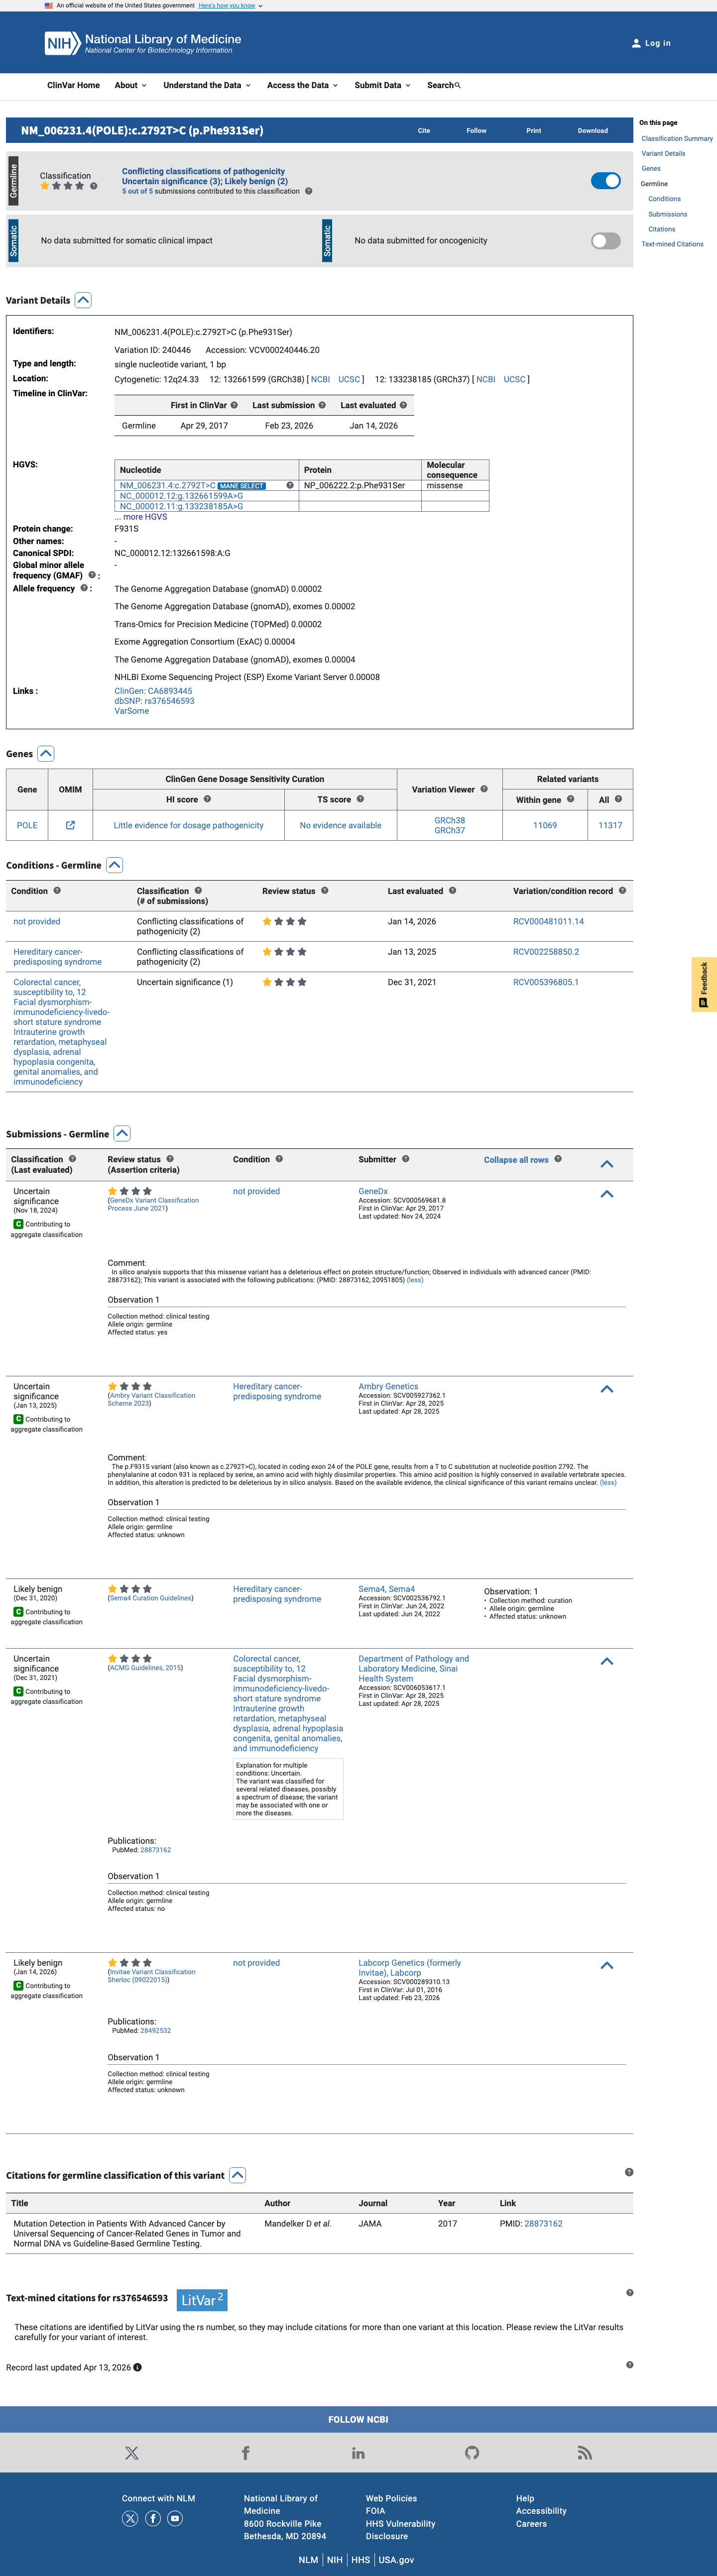
PP5 Not met: ClinVar VCV000240446 has zero expert-panel submissions (1-star, conflicting; 3 uncertain, 1 likely benign), so no expert-panel pathogenic assertion supports PP5.
Benign
BA1 Not met: the highest gnomAD subpopulation frequency is 0.00096432 (Ashkenazi Jewish, v2.1), far below the 0.05 BA1 threshold.
BS1 Not met: the highest gnomAD subpopulation frequency is 0.001014 (Ashkenazi Jewish, v2.1 non-cancer), below the 0.01 BS1 threshold.
BS2 Not met: gnomAD reports 0 homozygotes in v2.1, v4.1 and both non-cancer subsets, and the heterozygous carriers are unphenotyped.
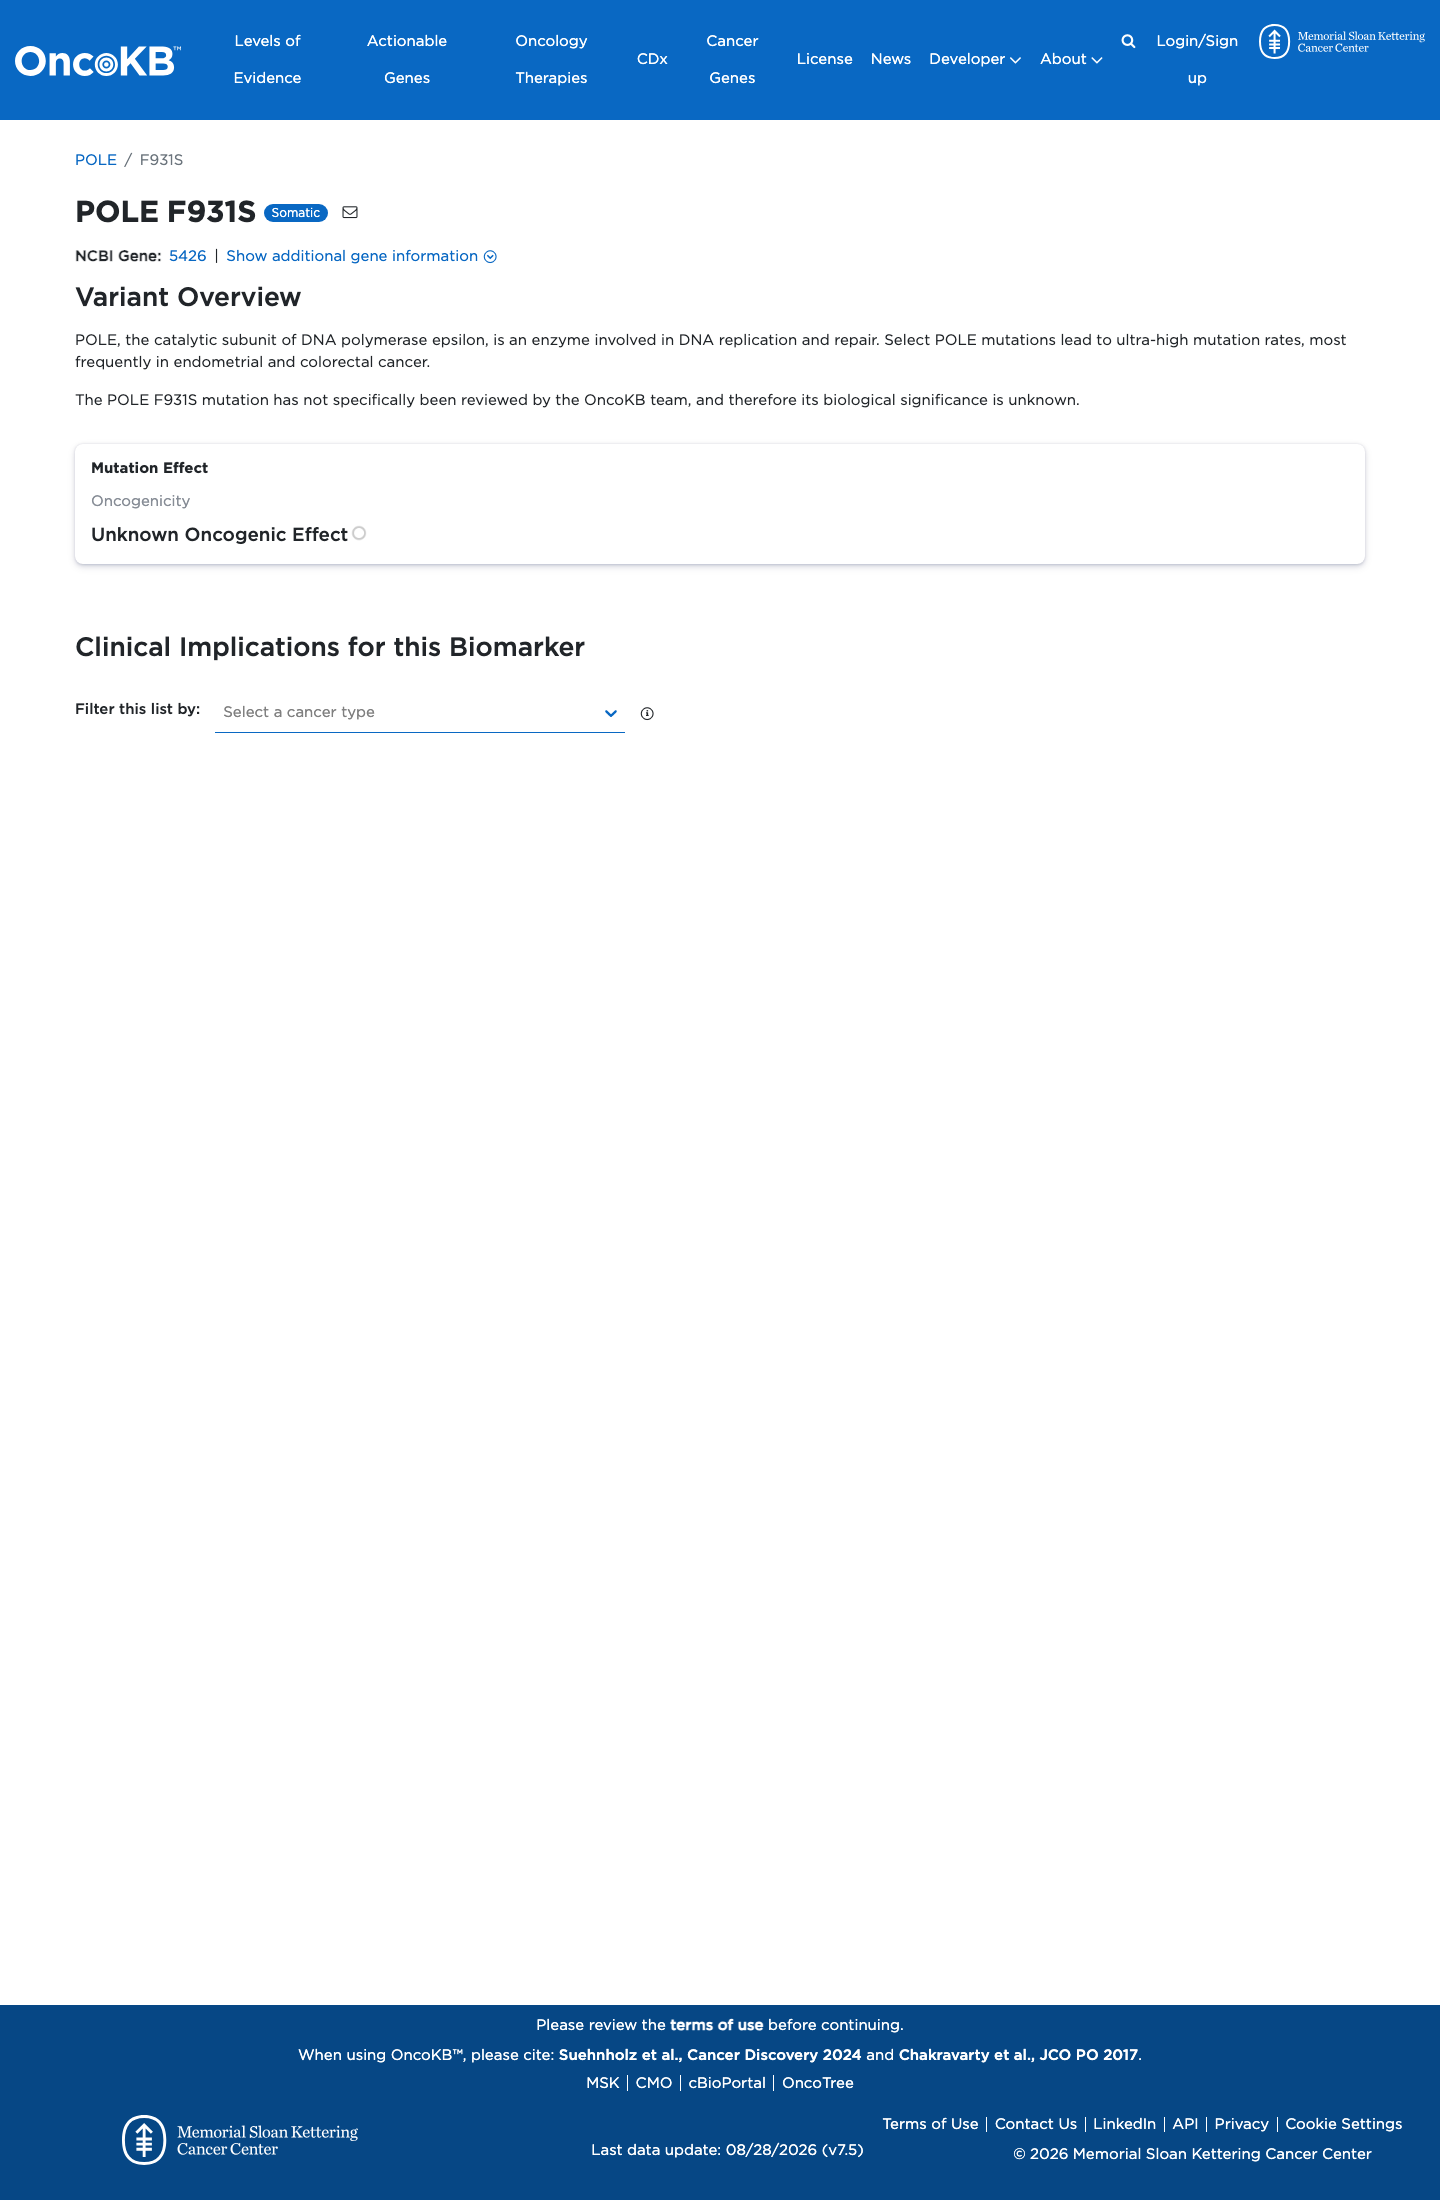
BS3 Not assessed: no functional data exist for p.Phe931Ser, and OncoKB records unknown oncogenic effect with no variant-specific evidence.
BS4 Not assessed: no family members with or without the variant are reported, so non-segregation cannot be evaluated.
BP1 Not met: POLE disease is driven by exonuclease-domain missense substitutions, not by truncating variants, so the premise of BP1 does not hold.
BP2 Not assessed: no cis/trans phase data exist linking c.2792T>C to any pathogenic POLE variant, so BP2 cannot be evaluated.
BP4 Not met: REVEL 0.749 is far above all BP4 thresholds (supporting cutoff <=0.29), so the missense computational evidence does not indicate a benign effect.
BP5 Not assessed: no case-level genotype or co-occurring alternate genetic cause was captured, so BP5 cannot be evaluated.
BP6 Not met: ClinVar VCV000240446 has no expert-panel submission (1-star, conflicting; 3 uncertain, 1 likely benign), so no expert-panel benign assertion supports BP6.
N/A · 4 PVS1 · PM4 · BP3 · BP7
Research & evidence
Population frequency
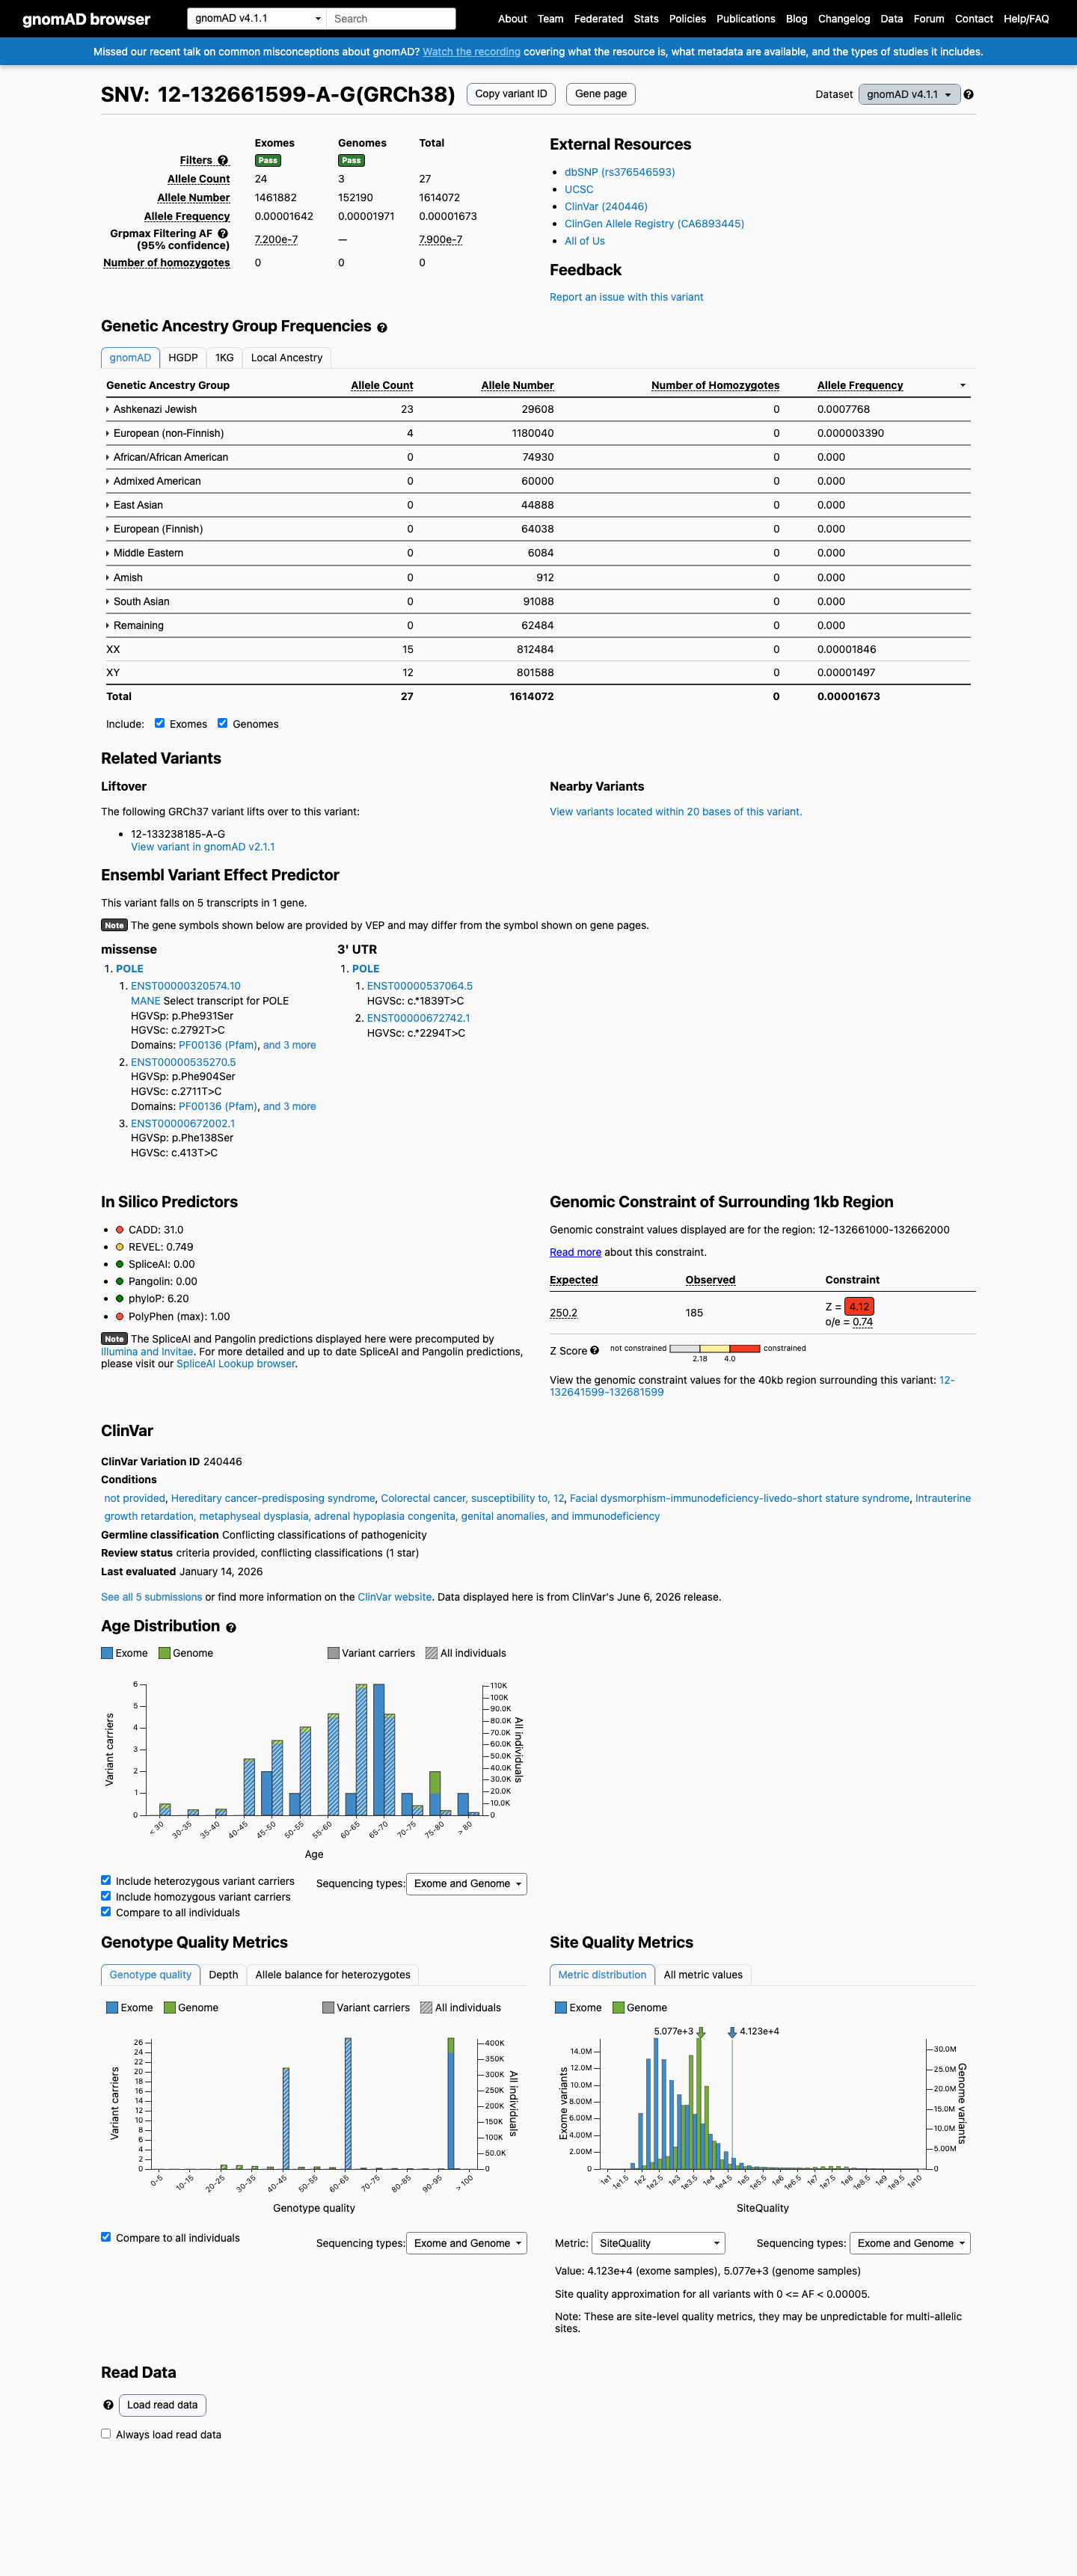
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 1.67279e-05; MAF= 0.00167%, 27/1614072 alleles, homozygotes = 0) and has highest observed frequency in the Ashkenazi Jewish population (AF= 0.000776817; MAF= 0.07768%, 23/29608 alleles, homozygotes = 0); grpmax FAF= 7.9e-07.
v2.1
This variant is present in gnomAD v2.1 (AF= 3.88871e-05; MAF= 0.00389%, 11/282870 alleles, homozygotes = 0) and has highest observed frequency in the Ashkenazi Jewish population (AF= 0.00096432; MAF= 0.09643%, 10/10370 alleles, homozygotes = 0).
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.0017% · 27 / 1,614,072
0 hom · FAF 7.9e-05%
Ashkenazi Jewish
23 / 29,608
0.078%
European (non-Finnish)
4 / 1,180,040
0.00034%
+ 8 not observed (Remaining individuals, Admixed American, European (Finnish), Amish, East Asian, Middle Eastern, South Asian, African/African American)
gnomAD v2.1
0.0039% · 11 / 282,870
0 hom
Ashkenazi Jewish
10 / 10,370
0.096%
European (non-Finnish)
1 / 129,190
0.00077%
+ 6 not observed (African/African American, Admixed American, East Asian, European (Finnish), Remaining individuals, South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Uncertain significance (3 clinical laboratories) and as Likely benign (1 clinical laboratory). (ClinVarID = 240446)
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.01). REVEL score = 0.749. BayesDel score = 0.145561.
Functional / OncoKB screenshot
Functional Unknown Oncogenic Effect
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. POLE, the catalytic subunit of DNA polymerase epsilon, is an enzyme involved in DNA replication and repair. Select POLE mutations lead to ultra-high m
OncoKB ↗
COSMIC screenshot
COSMIC
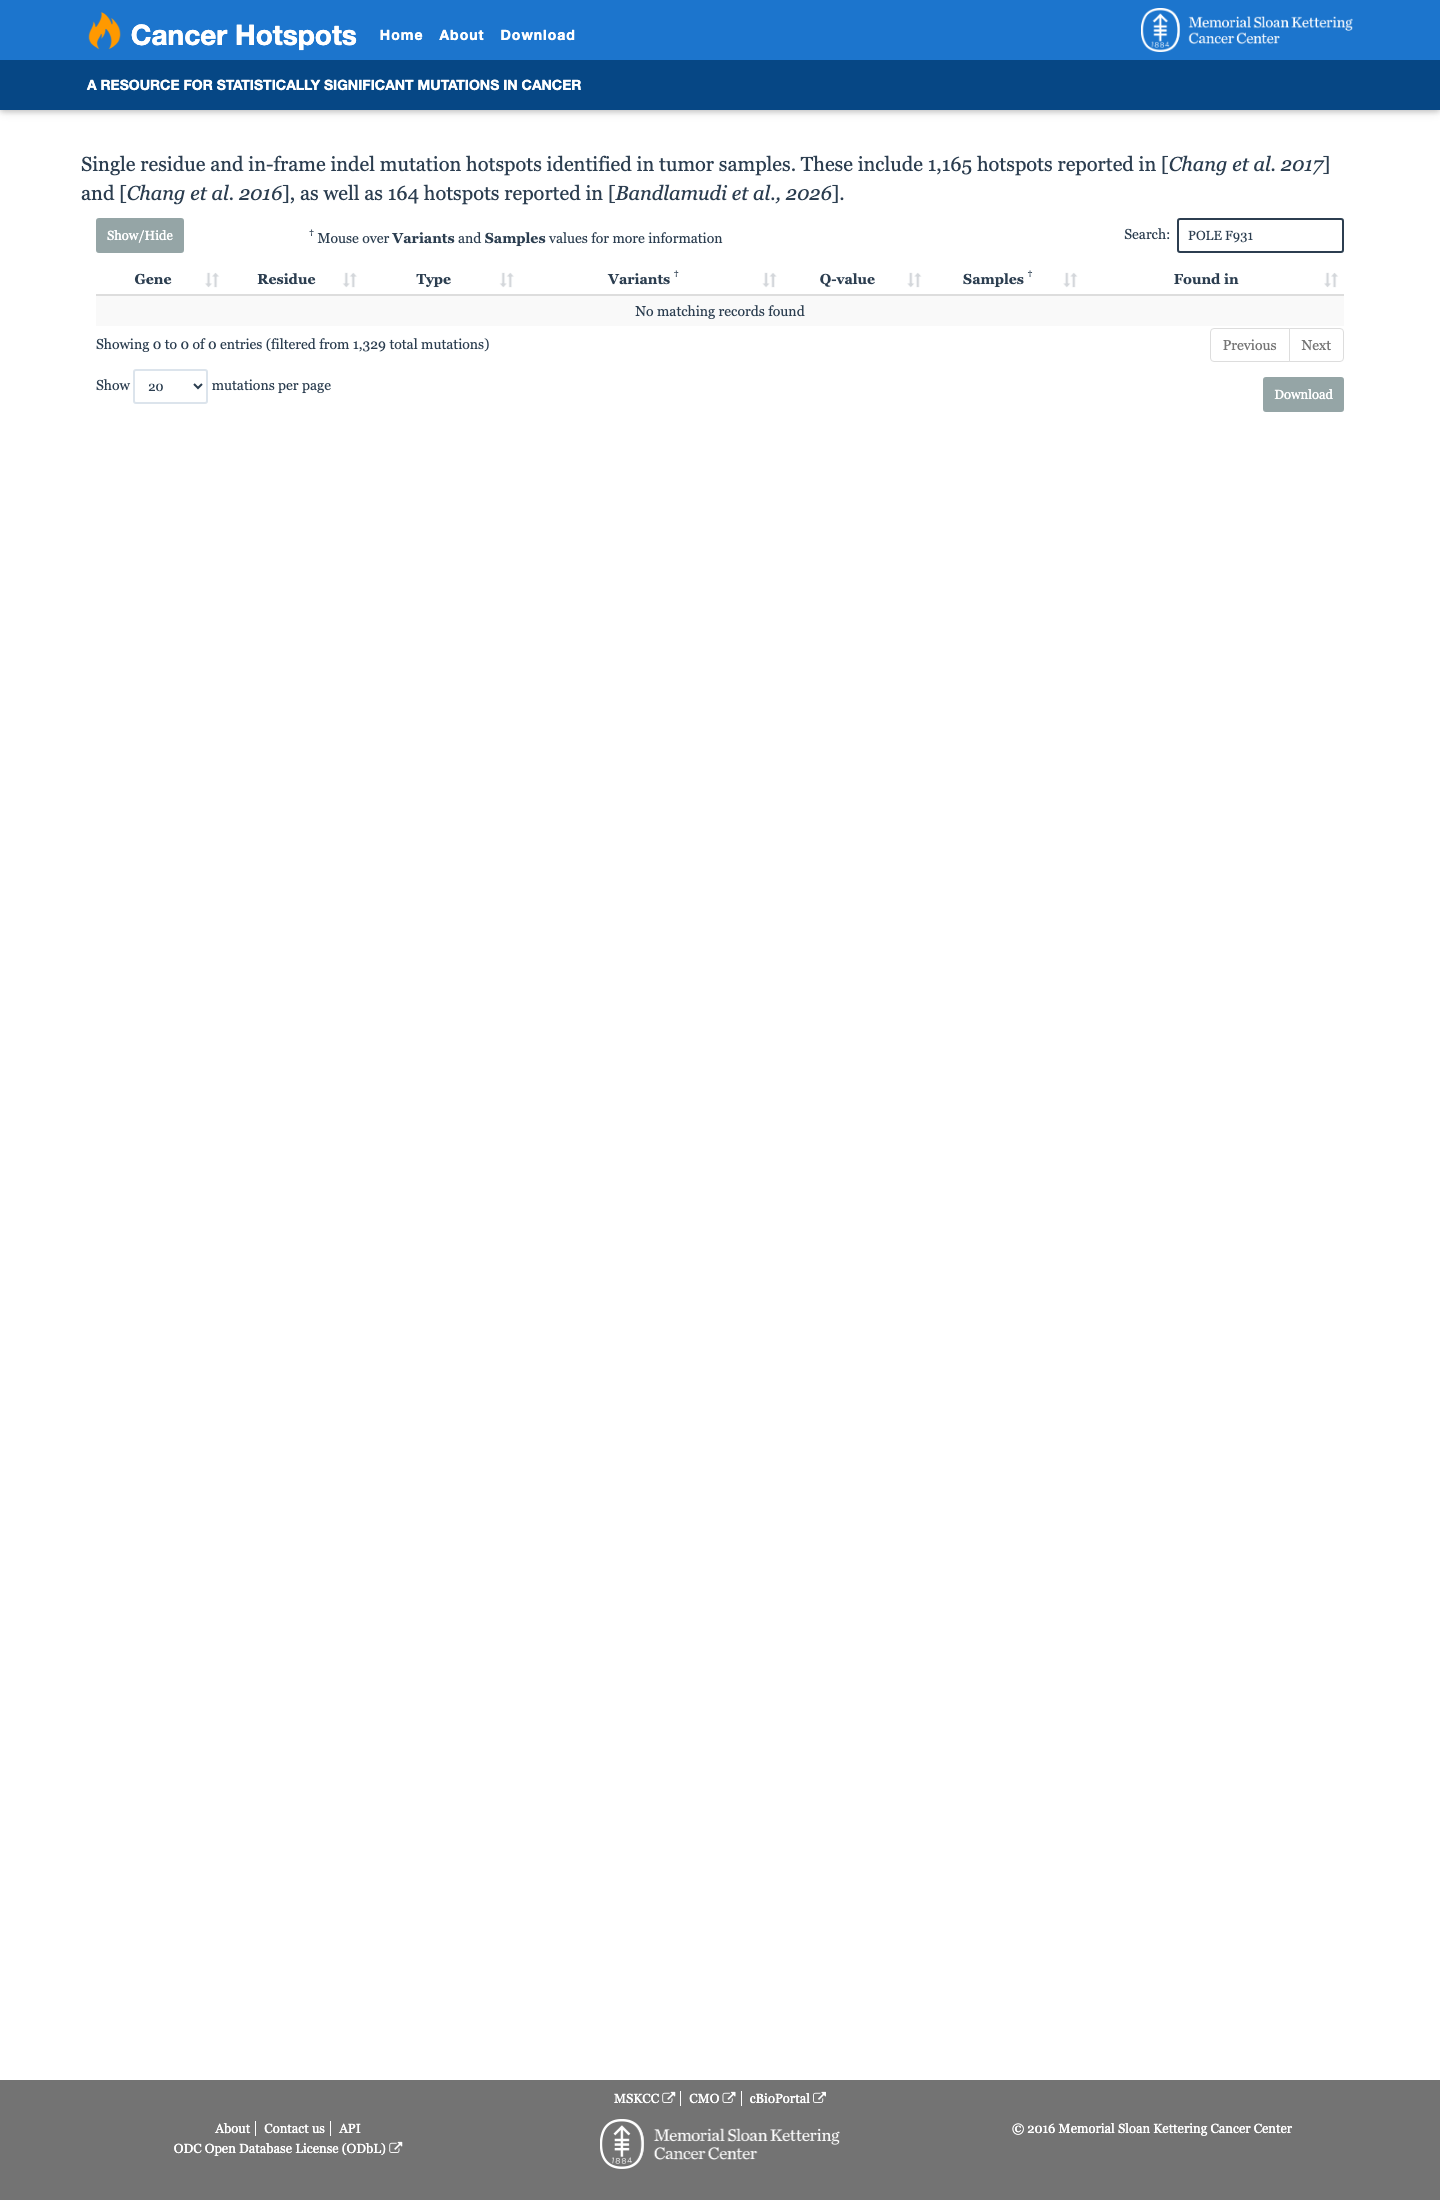
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
1papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 3 further PMIDs triaged but not cited — see Sources & references.
Rule & framework references · cited for criterion definitions, not variant evidence
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 3 PMIDs not cited in assessment
25394175 ↗ A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment. CLINVAR
28873162 ↗ Mutation Detection in Patients With Advanced Cancer by Universal Sequencing of Cancer-Related Genes in Tumor and Normal DNA vs Guideline-Based Germline Testing. CLINVAR
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification criteria. CLINVAR