Back
NM_006231.4:c.547G>A
p.Ala183Thr · POLE
León-Castillo et al. 2020 custom POLE framework · vleon-castillo-2020-custom-framework-v1
0%
complete
Final classification
VUS
PM2PP2BP4
POLE
c.547G>A
p.Ala183Thr
missense · exon 6

POLE encodes the catalytic subunit of DNA polymerase epsilon, the enzyme that replicates the leading strand of DNA during cell division and participates in DNA repair. It contains a proofreading domain that corrects replication errors, keeping the accumulation of mutations in check. Germline mutations in POLE cause polyposis and predispose to colorectal cancer, and are also linked to a rare syndrome of facial dysmorphism, immunodeficiency, livedo, and short stature. Somatic mutations, particularly in the proofreading domain, occur in colorectal and endometrial cancers, where they drive an ultra-mutated tumor phenotype and are associated with better responses to immune checkpoint inhibitors.

This variant

Ala183 lies N-terminal to and outside POLE's exonuclease proofreading domain (residues 268-471), the domain whose heterozygous missense defects cause polymerase proofreading-associated polyposis and colorectal cancer predisposition, so this substitution is not currently supported as a POLE-related cancer allele.

Transcript
NM_006231.4
HGVS · transcript:coding
NM_006231.4:c.547G>A
GRCh38
chr12:132679528 C>T
GRCh37
chr12:133256114 C>T
VUS: PM2 (supporting) and PP2 (supporting) give no pathogenic rule, and the single criterion BP4 (moderate) gives no benign rule.
Classification rationale
PM2PP2 BP4 VUS
POLE c.547G>A missense · exon 6

VUS: PM2 (supporting) is met because gnomAD v4.1 frequency 0.0000384 (62/1,613,808 alleles, no homozygotes) is below the 0.0001 rarity threshold. VUS: PP2 (supporting) is met because POLE germline disease is caused by missense proofreading-domain substitutions and this is a missense change. VUS: BP4 (moderate) is met because REVEL 0.037 falls inside the moderate benign band (<=0.183).

PM2 + PP2 + BP4 VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_006231.4 · variants mapped to exon structure
POLE NM_006231.4
Fetching transcript structure from UCSC…
Applied criteria · 3 applied · 21 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 3
Strength Supporting Moderate Strong Very strong
PM2 supporting Pathogenic
Met at Supporting: gnomAD v4.1 frequency 0.0000384 (62/1,613,808; grpmax FAF 0.0000378) is below the 0.0001 Supporting PM2 threshold.
gnomAD v4.1 (all-comers, GRCh38 chr12-132679528-C-T): total AF 3.84184e-05 (62/1,613,808 alleles, homozygotes 0), grpmax FAF 3.775e-05; exome 4.03695e-05 (59/1,461,500); genome 1.96969e-05 (3/152,308); highest ancestry European (non-Finnish) 4.83167e-05 (57/1,179,716). All values below 0.0001.gnomAD v2.1 (all-comers, GRCh37 12-133256114-C-T): total/exome AF 7.9552e-06 (2/251,408 alleles, homozygotes 0), grpmax FAF 2.92e-06; highest ancestry European (non-Finnish) 1.75889e-05 (2/113,708). Below 0.0001.Non-cancer sensitivity check: gnomAD v2.1 non-cancer exomes AF 8.44338e-06 (2/236,872 alleles, homozygotes 0); gnomAD v3.1 non-cancer genomes AF 2.02782e-05 (3/147,942 alleles, homozygotes 0). Both below 0.0001, so the source choice does not change the verdict.
PP2 supporting review Pathogenic
Met at supporting: missense exonuclease-domain substitutions are the established POLE disease mechanism, though no gnomAD missense-constraint metric was available to confirm rare benign missense variation.
PP2 definition (PMID:25741868): missense variant in a gene that has a low rate of benign missense variation and in which missense variants are a common mechanism of disease.OncoKB gene-level curation: 'Germline heterozygous loss-of-function mutations in the exonuclease domain of POLE cause polyposis and predispose individuals to colorectal cancer' - these germline alleles are missense proofreading-domain substitutions, establishing missense as a common disease mechanism for POLE.OncoKB background states that POLE 'contains an exonuclease "proofreading" domain' whose alteration results in accumulation of single-nucleotide variants and an ultramutated phenotype - a missense-driven mechanism.
BP4 moderate Benign
Met at moderate strength: REVEL 0.037 falls below the 0.183 moderate BP4 threshold.
The local POLE custom framework's BP4 rule requires the exact missense variant to appear in Supplementary Table S2 or S3 with REVEL class 'Likely benign' and >=4 benign in-silico results, and explicitly directs fallback to the generic in-silico workflow when the variant is absent from those tables.Direct search of the converted Supplementary Table S2 (PATH-250-323-s003.xlsx.txt) and Supplementary Table S3 (PATH-250-323-s004.xlsx.txt) found no entry for p.A183T, p.Ala183Thr or c.547G>A in either table.REVEL score 0.037 from the local REVEL v1.3 lookup (GRCh38 chr12:132679528 C>T), compared against the supplied ClinGen SVI REVEL benign calibration: BP4 supporting <=0.29, moderate <=0.183, strong <=0.016 (Pejaver et al. 2022, PMID:36413997).
Assessed · not applied · 11 not met · 10 not assessed
Pathogenic
PS1 Not met: the governing POLE pathogenic set is p.P286R, p.V411L, p.S297F, p.A456P and p.S459F, and ClinVar records no pathogenic p.Ala183Thr change.
PS2 Not assessed: no confirmed de novo occurrence for c.547G>A, since no parental testing or proband data exists in any available source.
PS3 Not assessed: no functional assay data exist for POLE p.Ala183Thr; the only retrieved functional claims are in-silico (REVEL 0.037), which cannot establish PS3.
PS4 Not met: A183T is absent from Supplementary Table S1 (combined endometrial-cancer recurrence count 0, not >=10), so the POLE custom PS4 rule cannot fire.
PM1 Not met: p.Ala183Thr lies outside the POLE exonuclease/proofreading domain, where the hotspots (p.P286R-p.S459F) and all framework domain variants are located.
PM3 Not assessed: no second POLE variant or phasing data exist, and gnomAD shows 2/251,408 alleles with zero homozygotes, so in-trans configuration is untested.
PM5 Not met: no pathogenic missense change at POLE residue 183 is reported, and same-residue candidate harvesting returned zero comparators.
PM6 Not assessed: no assumed de novo report for c.547G>A exists, as no proband, parental samples or family data were available.
PP1 Not assessed: no pedigree, affected relatives or meioses are reported for c.547G>A, so co-segregation cannot be evaluated.
PP3 Not met: REVEL 0.037 is far below the 0.644 supporting PP3 threshold, and no splice prediction was available.
PP4 Not assessed: no proband phenotype, HPO terms, or family history were captured, so phenotype specificity for POLE cannot be evaluated.
PP5 Not met: ClinVar 540667 has zero expert-panel submissions (1-star, conflicting: 3 uncertain, 1 likely benign), so no expert-panel pathogenic assertion supports PP5.
Benign
BA1 Not met: gnomAD v4.1 allele frequency 0.0000384 (62/1,613,808 alleles) is roughly 1,300-fold below the 0.05 stand-alone benign threshold.
BS1 Not met: the highest adequately powered population frequency, 0.0000483 in European (non-Finnish), is about 200-fold below the 0.01 BS1 threshold.
BS2 Not met: zero homozygotes in every gnomAD cohort, and POLE cancer predisposition is adult-onset and incompletely penetrant, failing BS2's fully-penetrant-early premise.
BS3 Not assessed: no functional assay of POLE p.Ala183Thr shows a non-damaging effect; the benign-leaning in-silico REVEL 0.037 supports BP4, not BS3.
BS4 Not assessed: no family with multiple affected members is reported for c.547G>A, so non-segregation cannot be demonstrated.
BP1 Not met: POLE disease is caused by missense proofreading-domain substitutions such as p.P286R and p.V411L, so BP1's truncating-only premise fails.
BP2 Not assessed: no data place this variant in cis or trans with a pathogenic POLE variant, and gnomAD reports zero homozygotes in every dataset.
BP5 Not assessed: no case-level genotype or co-occurring alternate genetic cause was captured, so BP5 cannot be evaluated.
BP6 Not met: ClinVar 540667 has no expert-panel submission, and the single Likely benign laboratory call is excluded from triggering BP6.
N/A · 4 PVS1 · PM4 · BP3 · BP7
Research & evidence
Population frequency
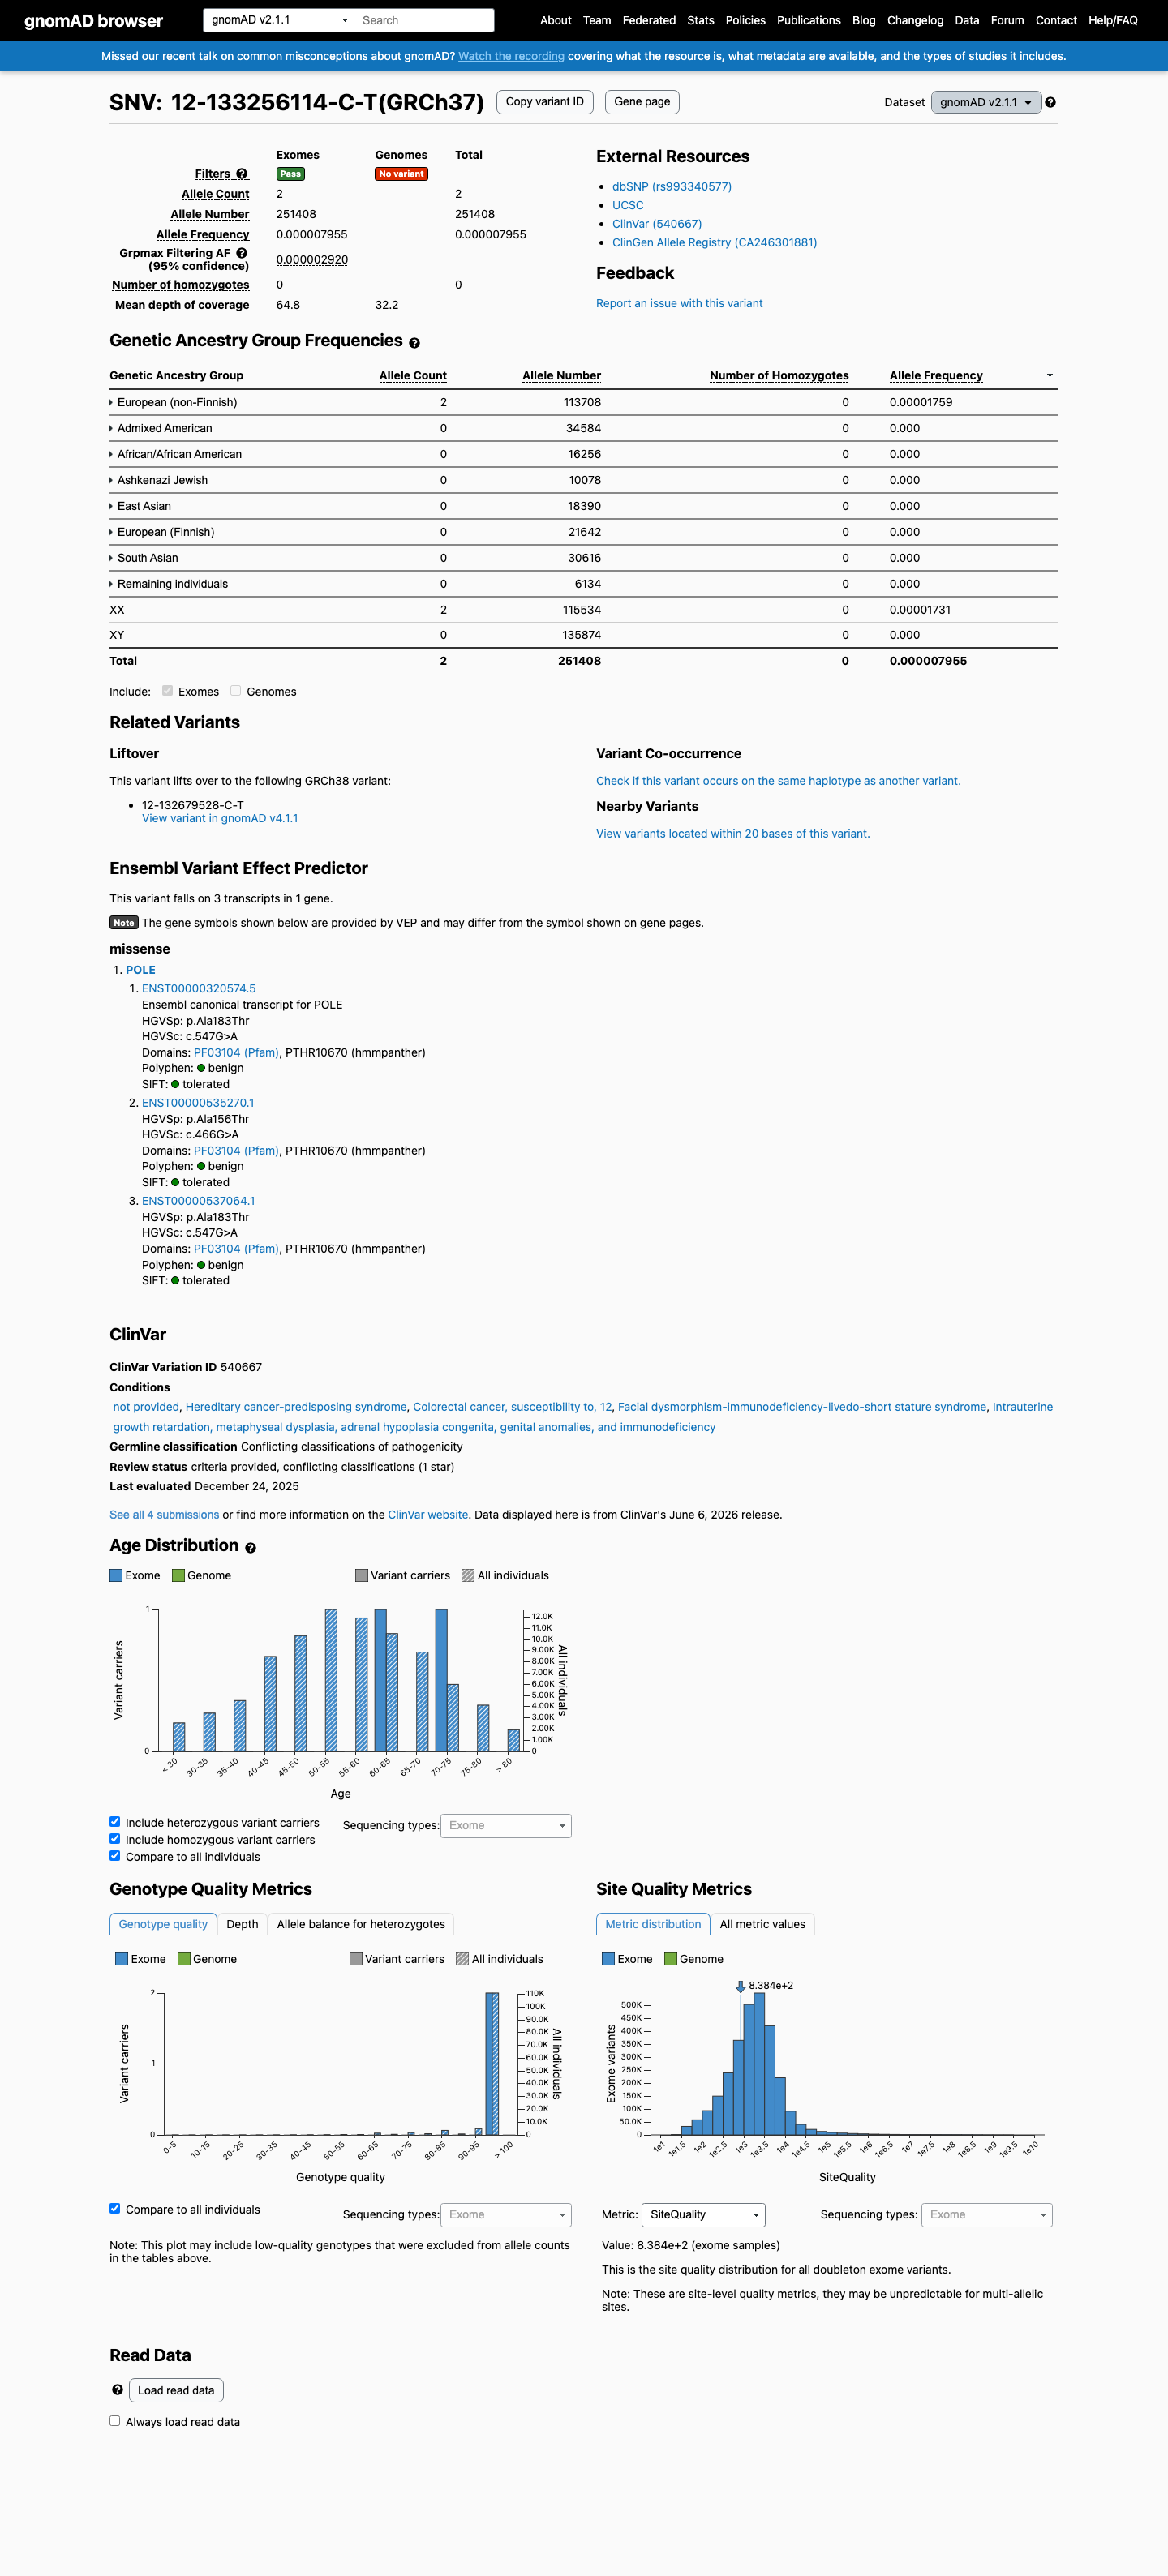
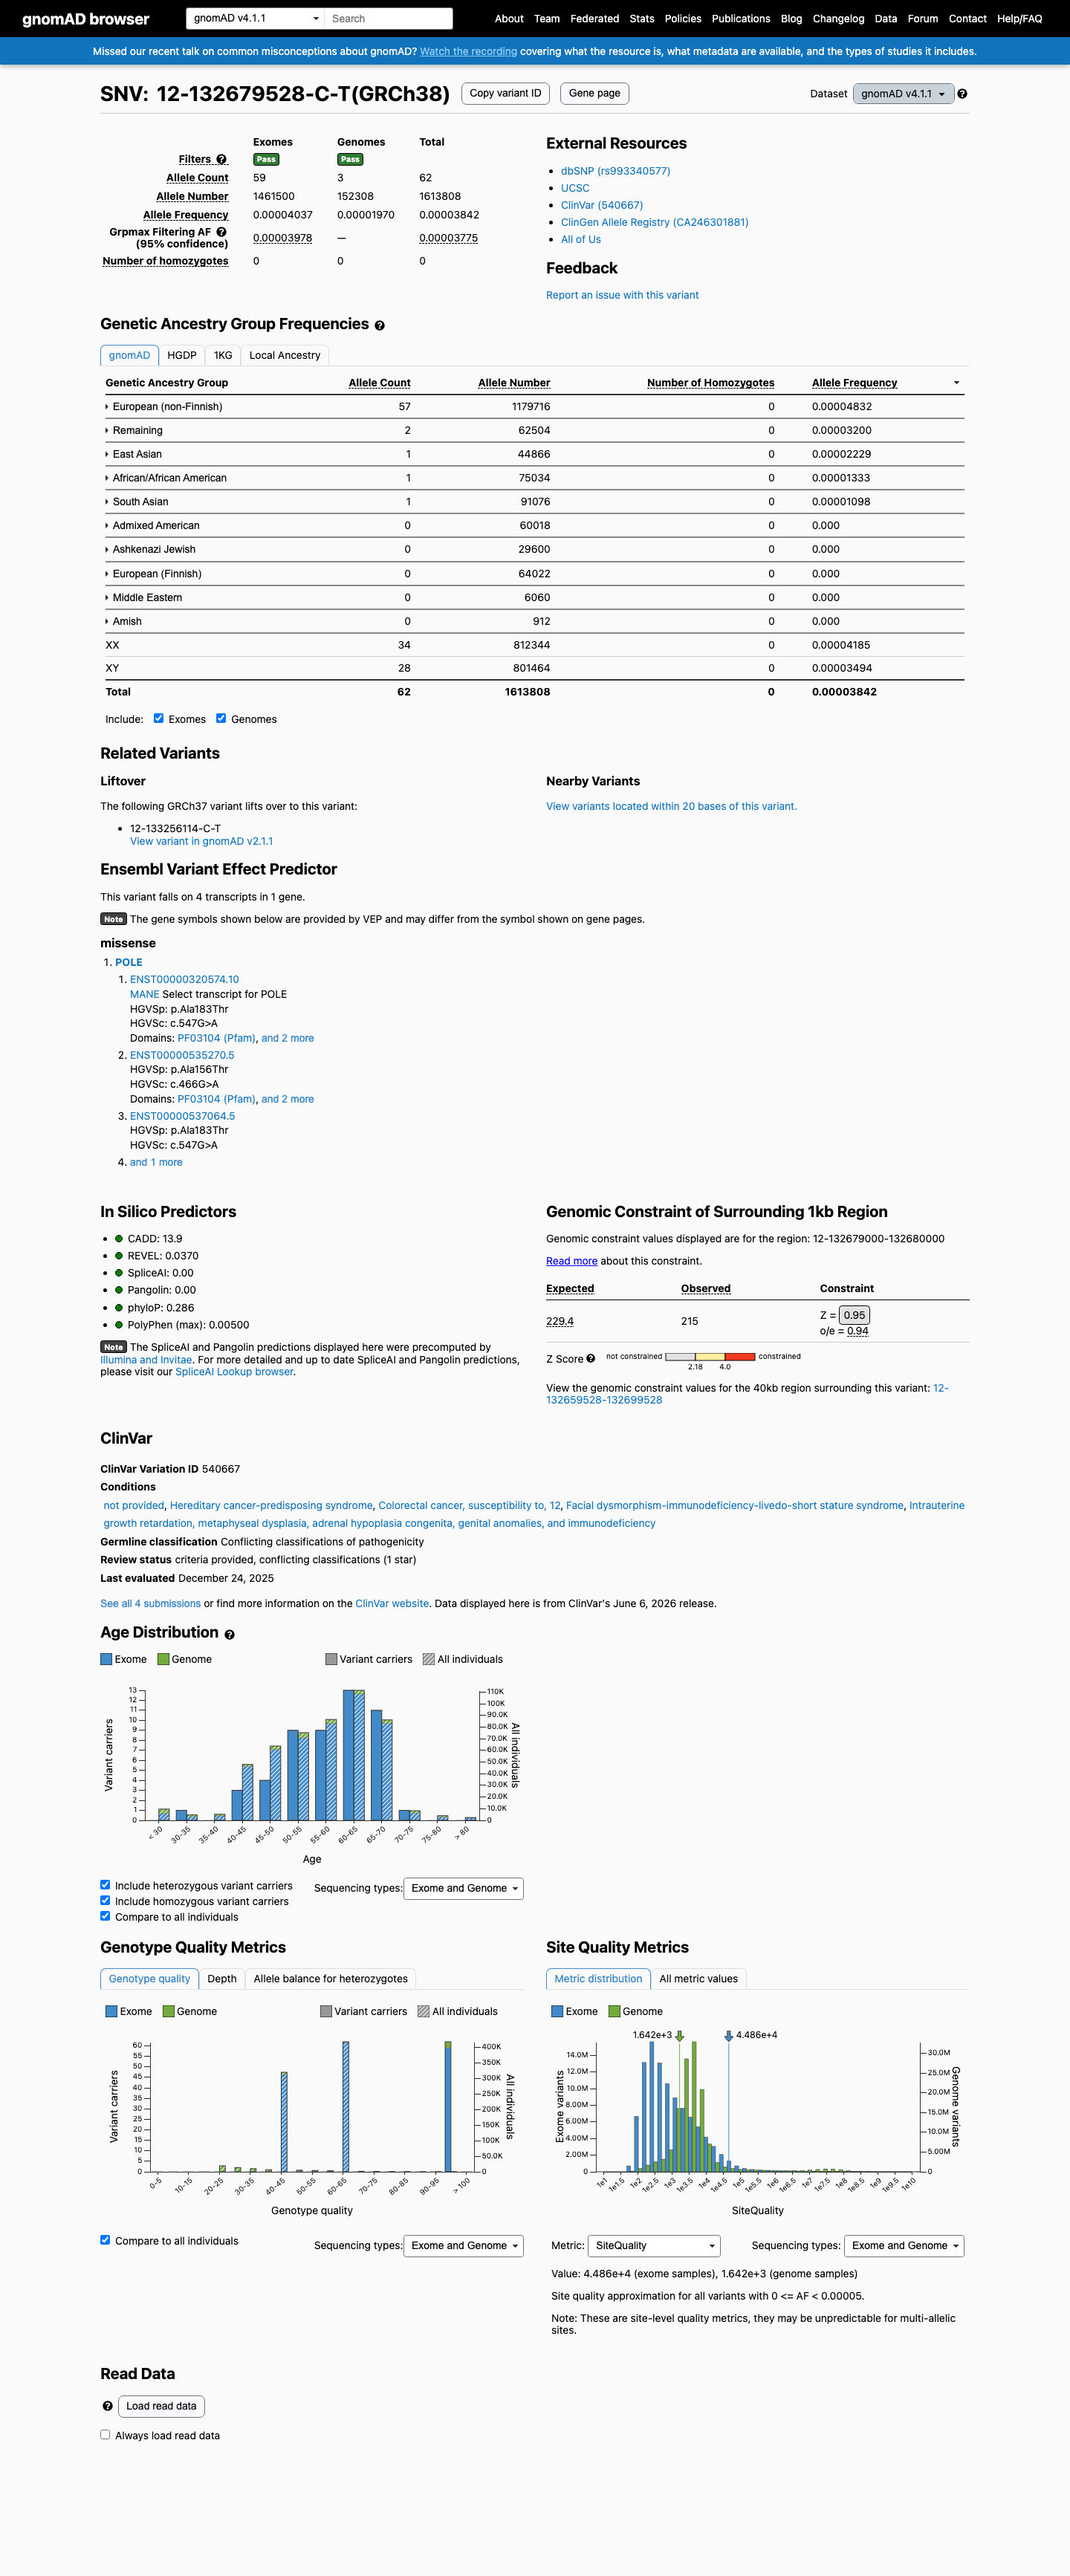
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 3.84184e-05; MAF= 0.00384%, 62/1613808 alleles, homozygotes = 0) and has highest observed frequency in the European (non-Finnish) population (AF= 4.83167e-05; MAF= 0.00483%, 57/1179716 alleles, homozygotes = 0); grpmax FAF= 3.775e-05.
v2.1
This variant is present in gnomAD v2.1 (AF= 7.9552e-06; MAF= 0.00080%, 2/251408 alleles, homozygotes = 0) and has highest observed frequency in the European (non-Finnish) population (AF= 1.75889e-05; MAF= 0.00176%, 2/113708 alleles, homozygotes = 0); grpmax FAF= 2.92e-06.
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.0038% · 62 / 1,613,808
0 hom · FAF 0.0038%
European (non-Finnish)
57 / 1,179,716
0.0048%
Remaining individuals
2 / 62,504
0.0032%
East Asian
1 / 44,866
0.0022%
African/African American
1 / 75,034
0.0013%
South Asian
1 / 91,076
0.0011%
+ 5 not observed (Admixed American, European (Finnish), Amish, Middle Eastern, Ashkenazi Jewish)
gnomAD v2.1
0.0008% · 2 / 251,408
0 hom · FAF 0.00029%
European (non-Finnish)
2 / 113,708
0.0018%
+ 7 not observed (African/African American, Admixed American, Ashkenazi Jewish, East Asian, European (Finnish), Remaining individuals, South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
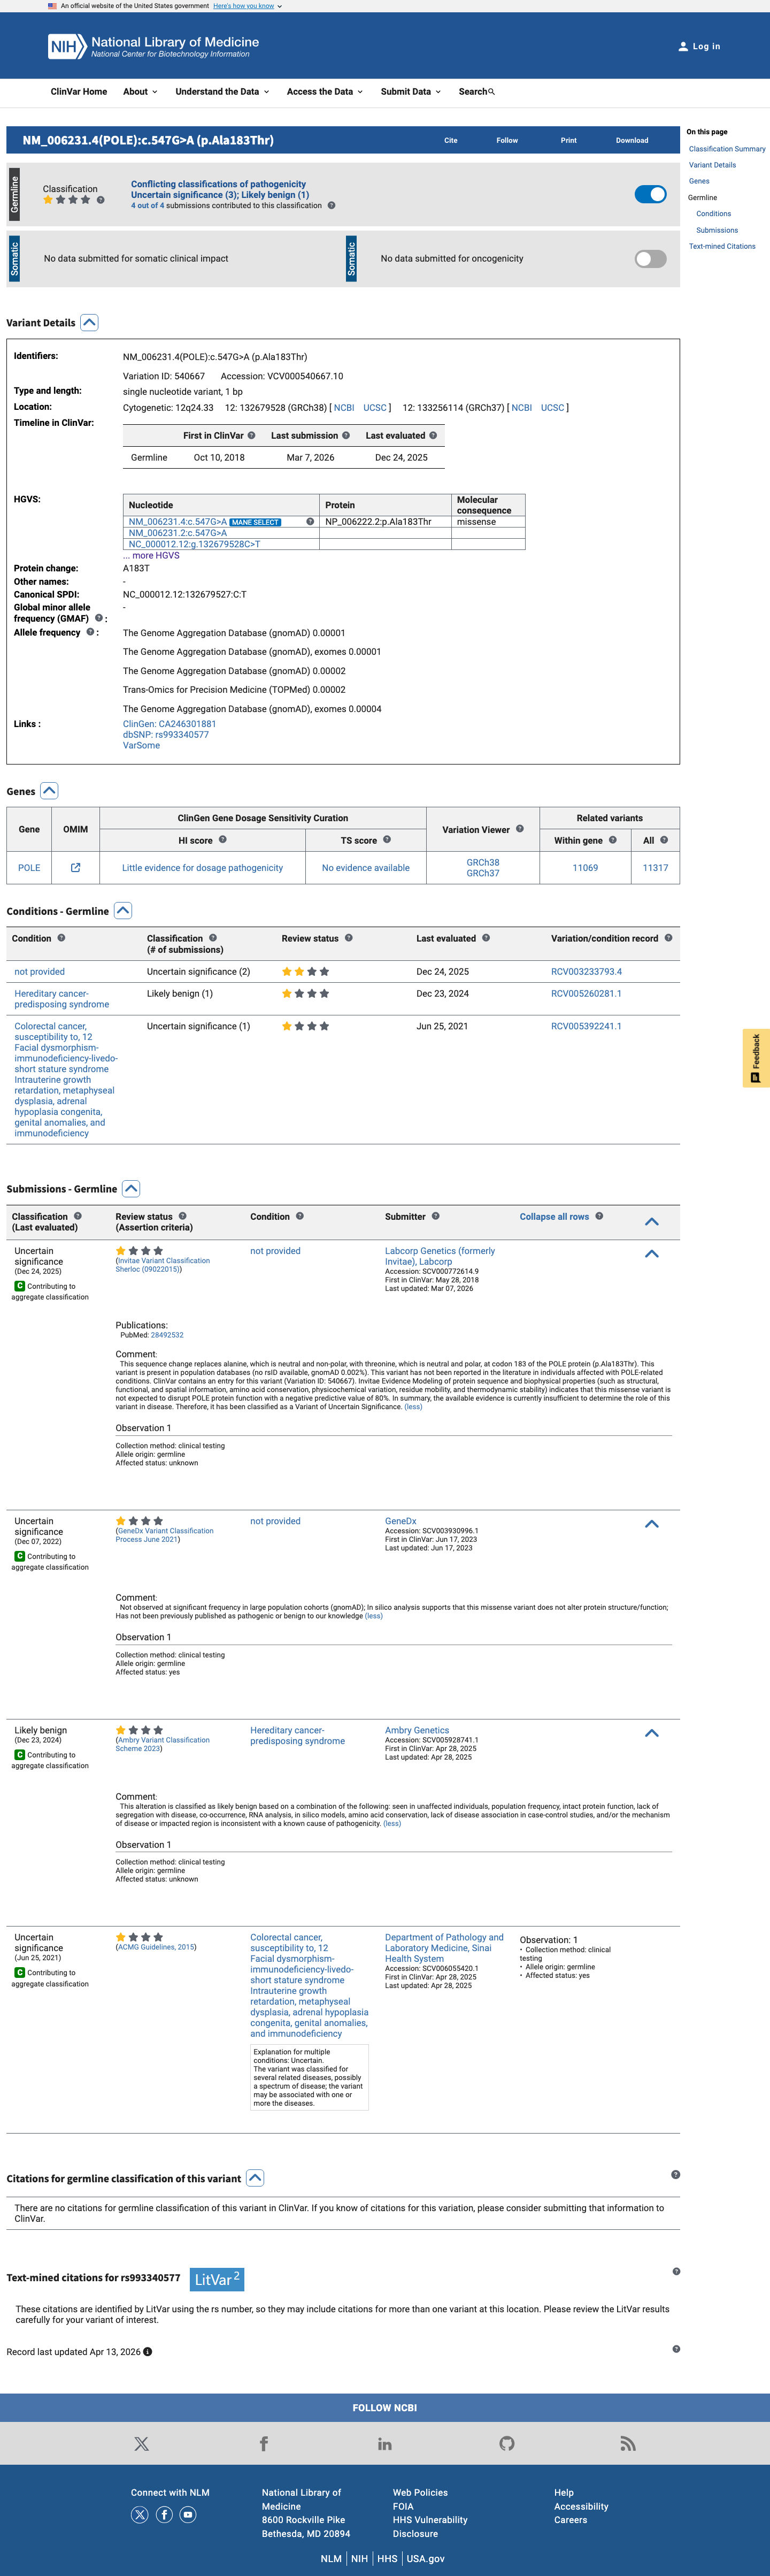
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Uncertain significance (3 clinical laboratories) and as Likely benign (1 clinical laboratory). (ClinVarID = 540667)
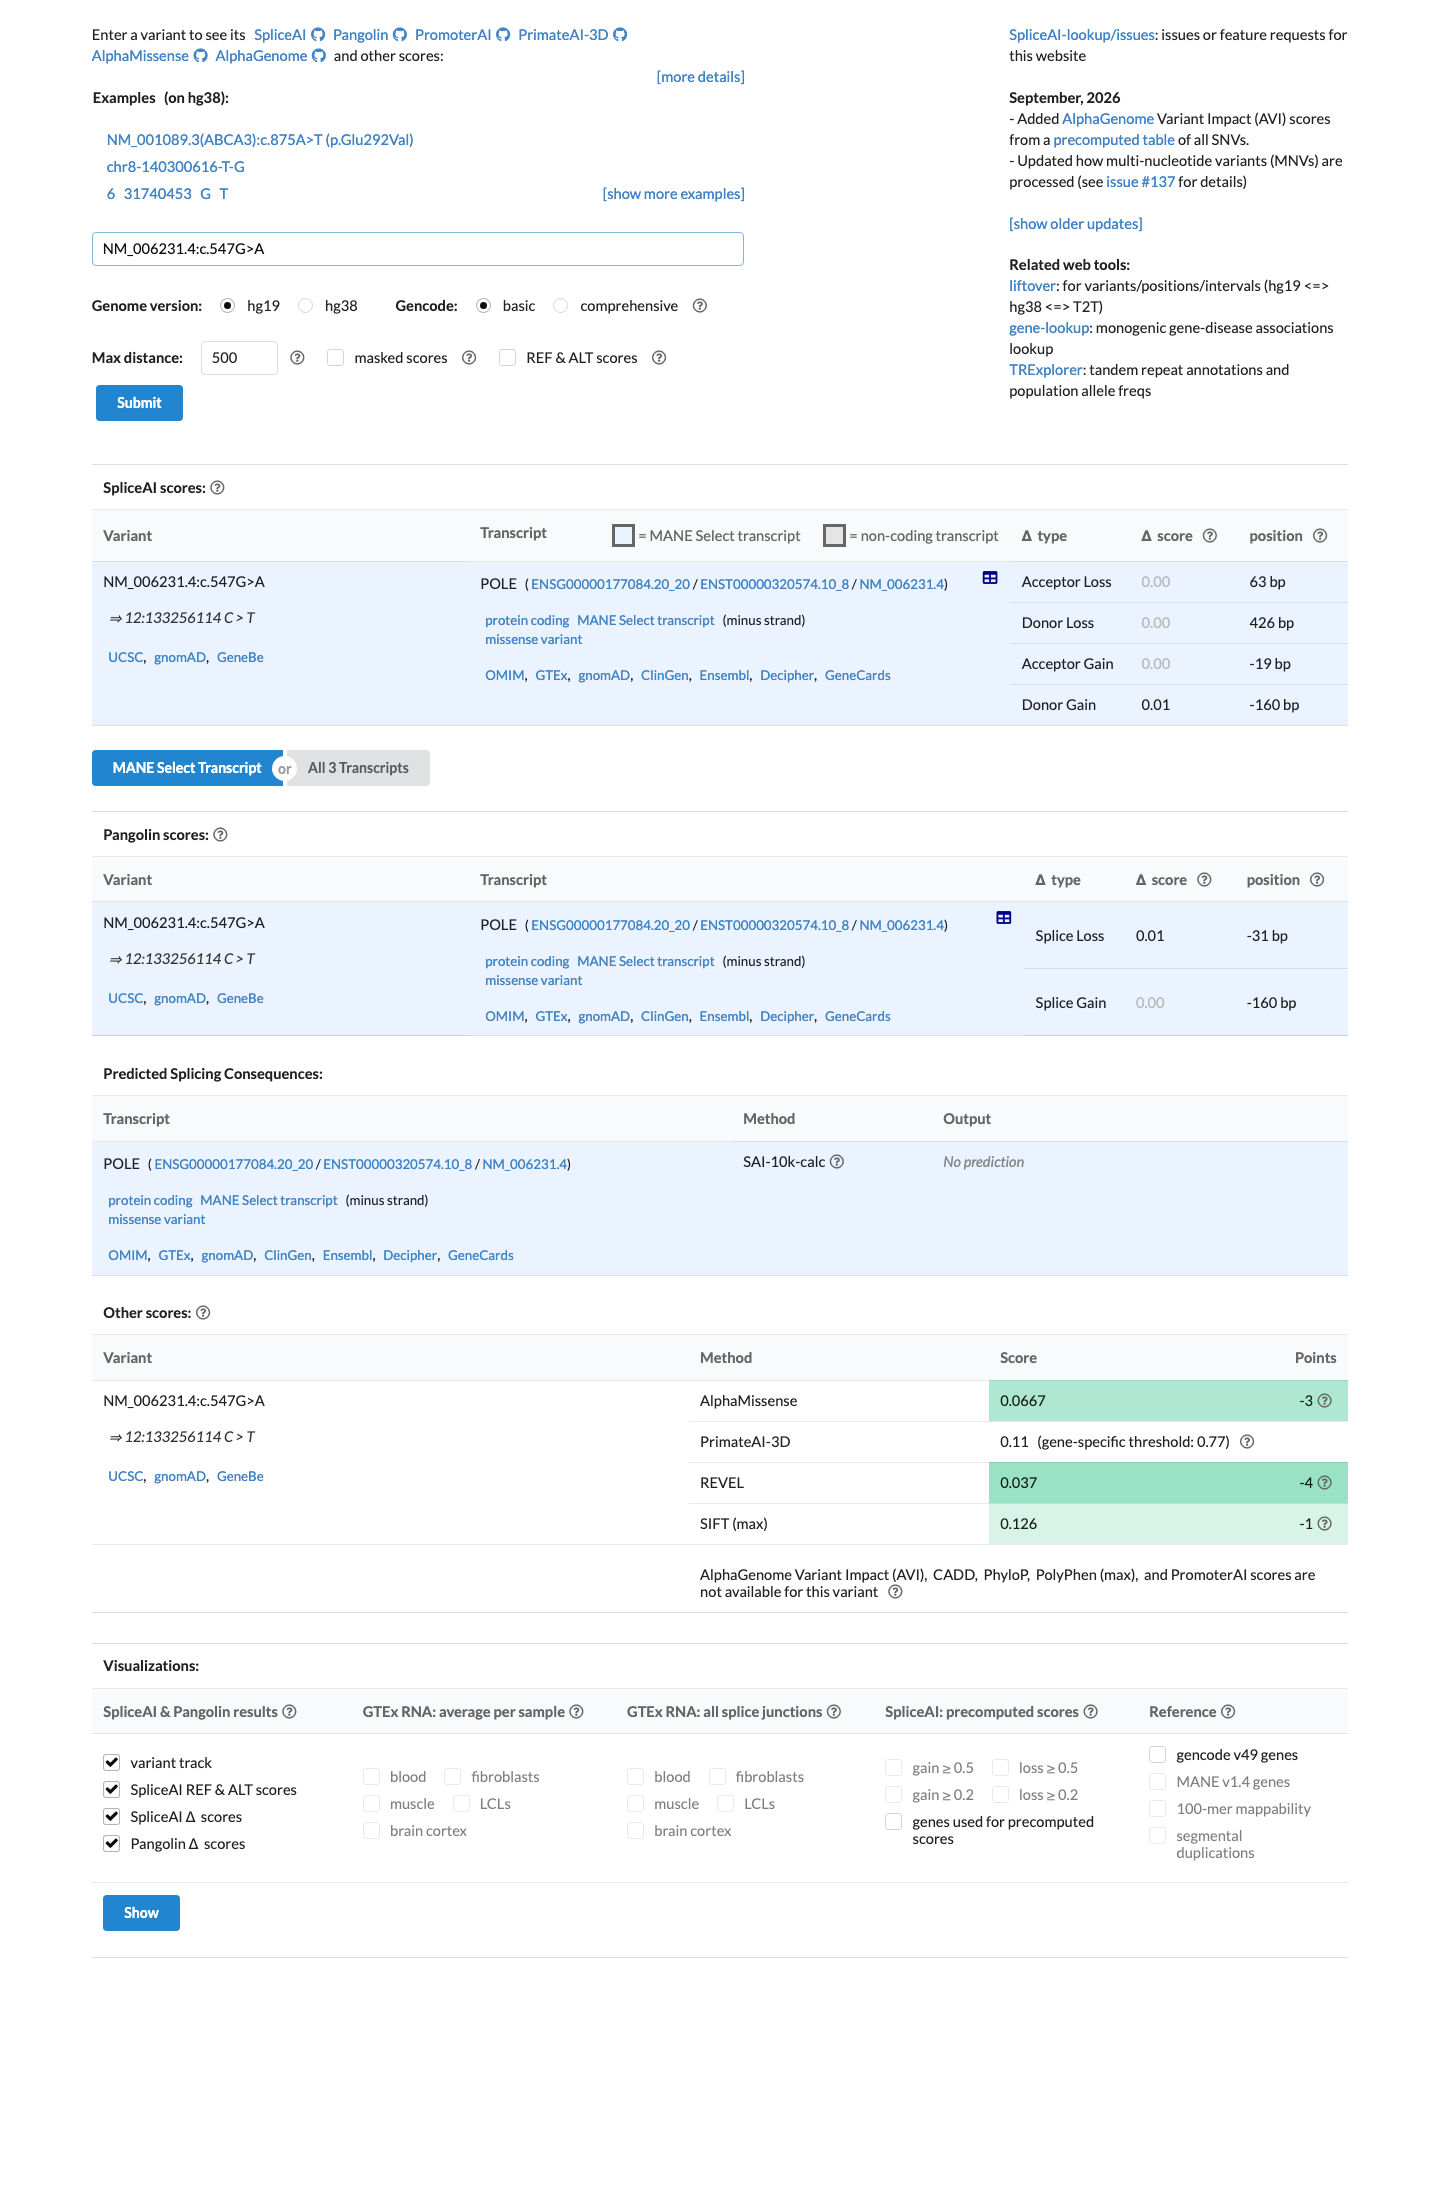
SpliceAI screenshot
In silico
SpliceAI returned NO scores for this variant, so no SpliceAI-based splice prediction is available. This is missing data, NOT evidence of absent splice impact: it must not be used to support BP4 or to argue against PP3/PVS1. Pangolin scores may be present but are not calibrated for PP3/BP4 here. REVEL score = 0.037. BayesDel score = -0.676197.
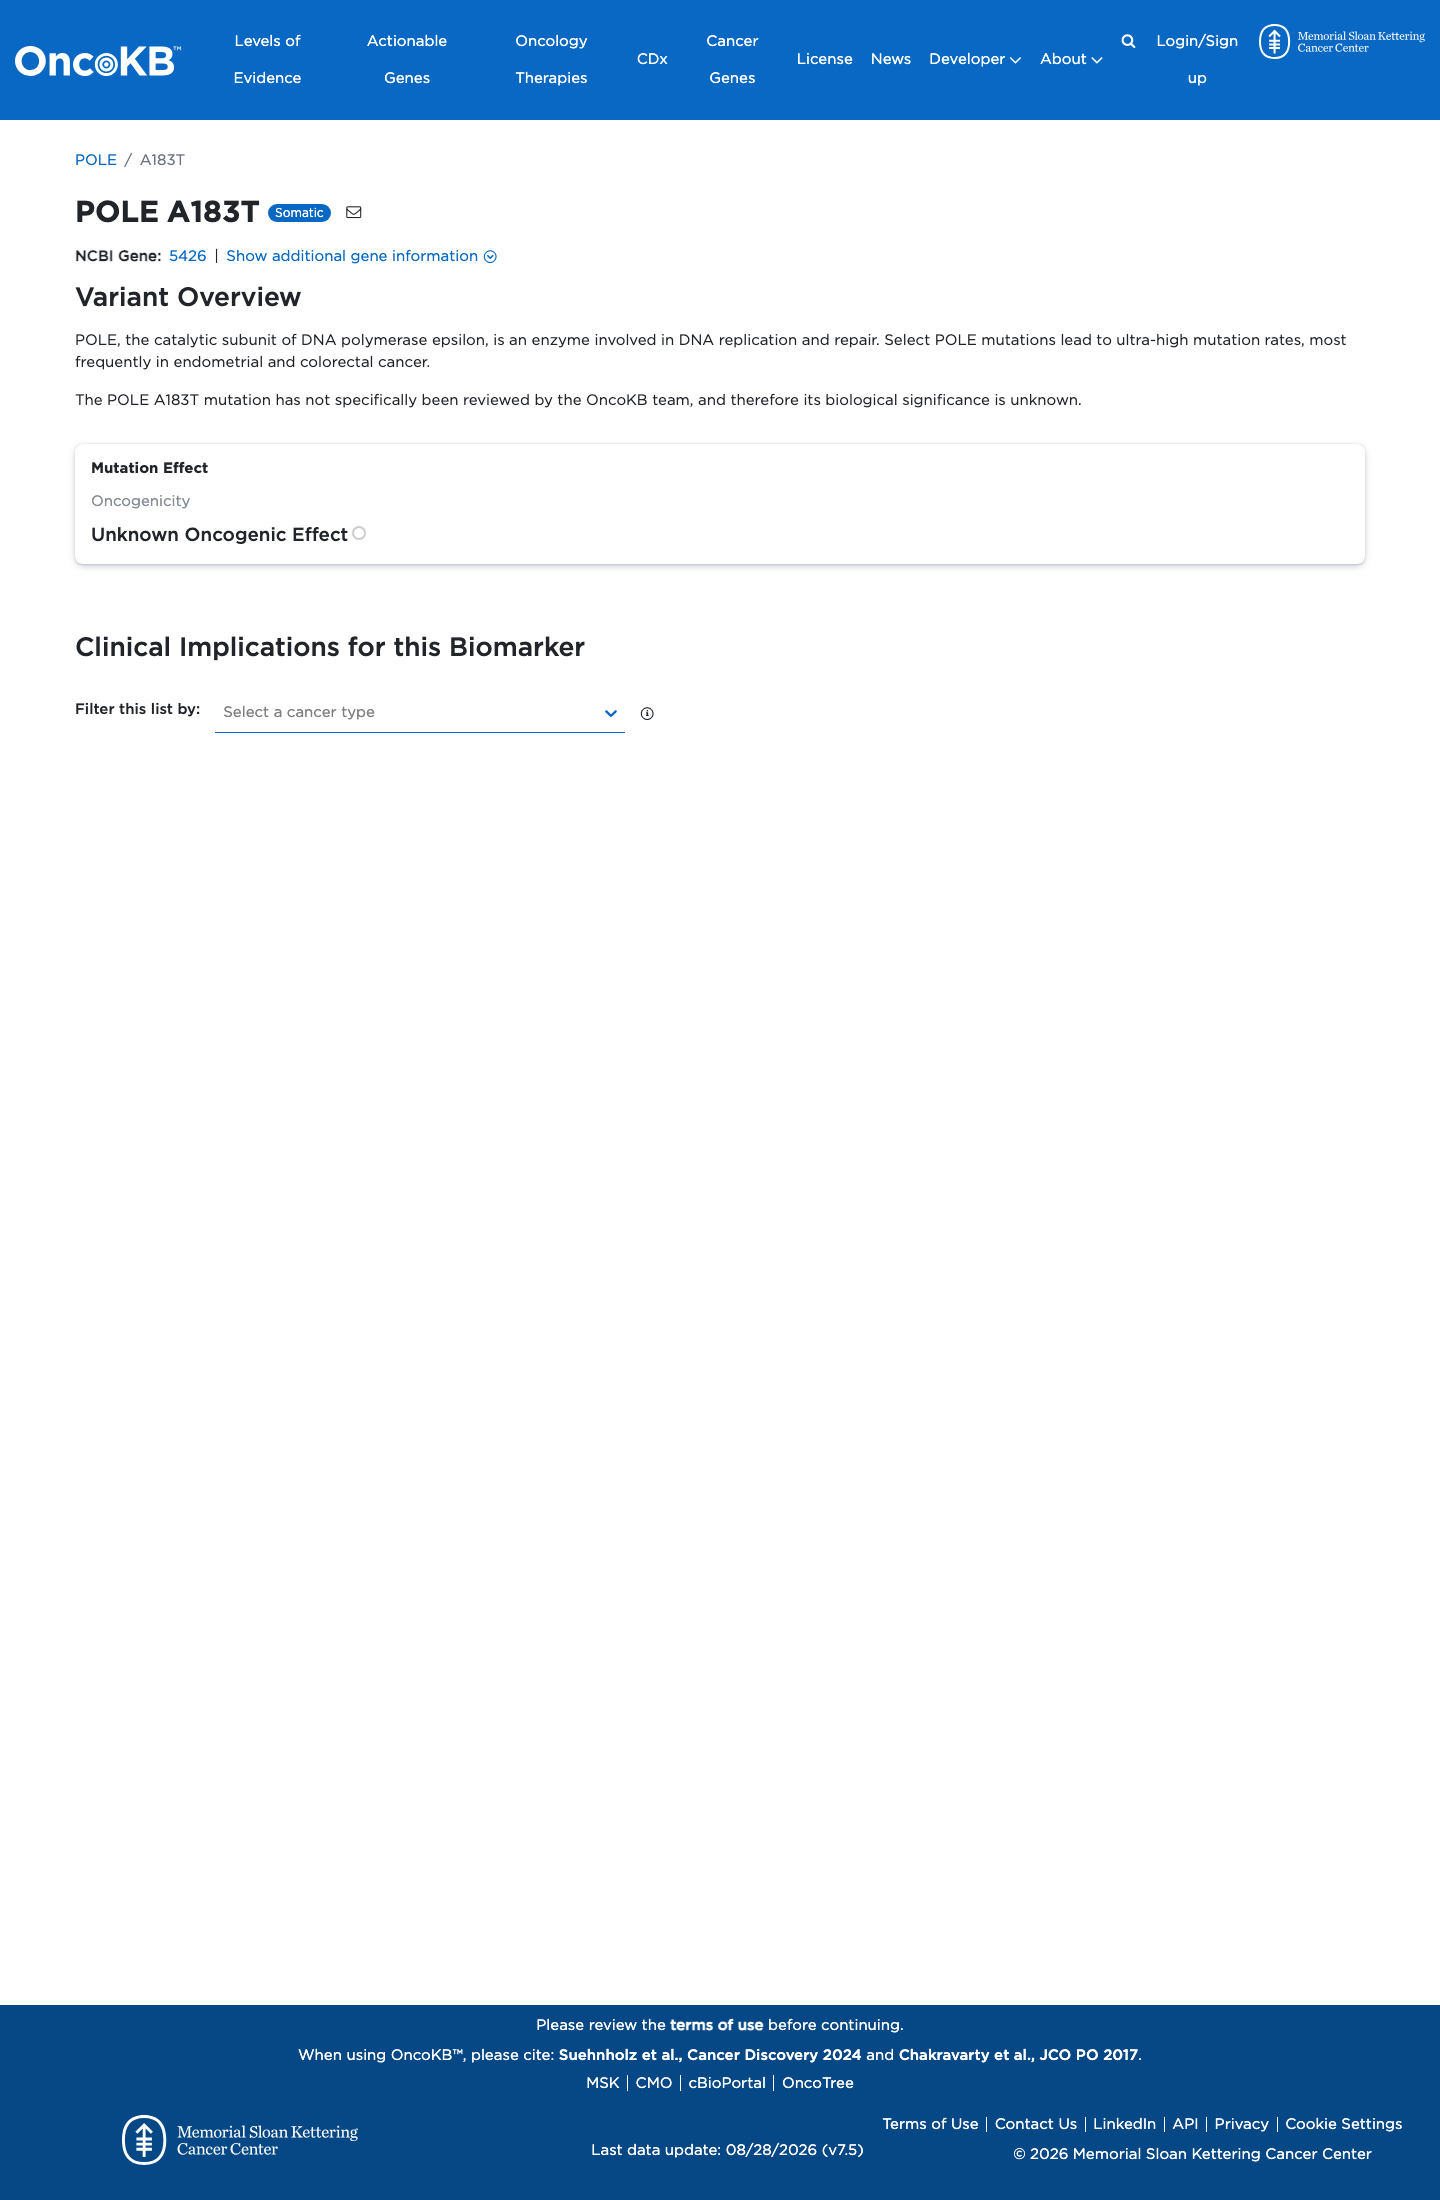
Functional / OncoKB screenshot
Functional Unknown Oncogenic Effect
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. POLE, the catalytic subunit of DNA polymerase epsilon, is an enzyme involved in DNA replication and repair. Select POLE mutations lead to ultra-high m
OncoKB ↗
COSMIC screenshot
COSMIC
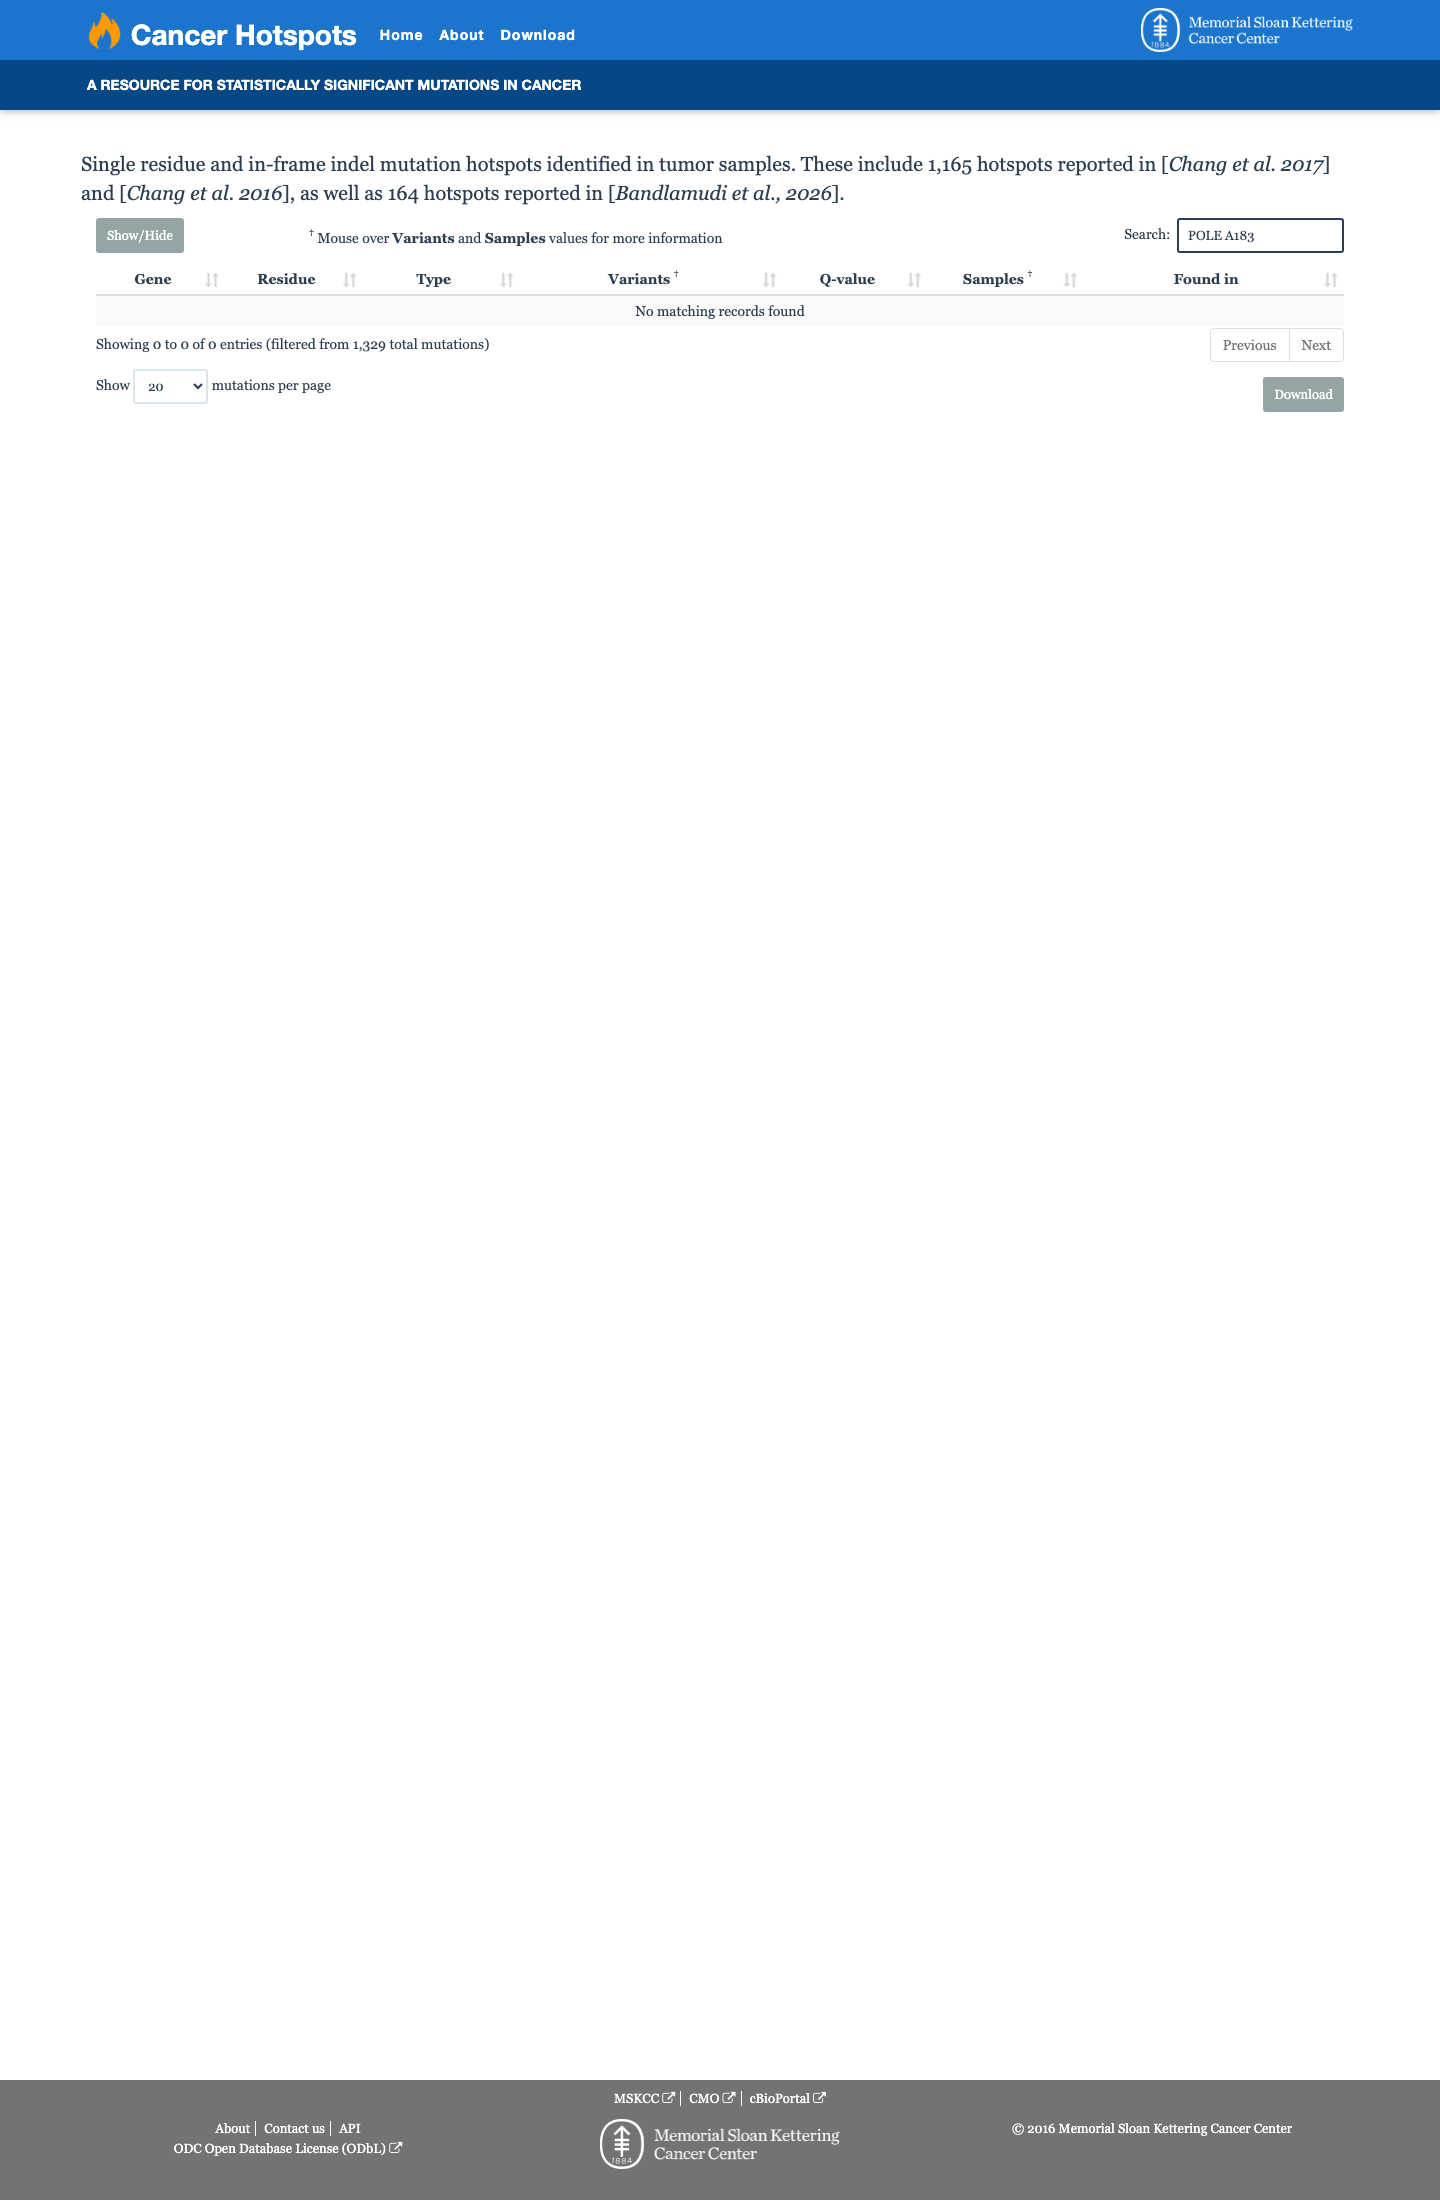
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
2papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why.
Rule & framework references · cited for criterion definitions, not variant evidence
25394175 ↗ A practice guideline from the American College of Medical Genetics and Genomics and the National Society of Genetic Counselors: referral indications for cancer predisposition assessment.
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots