Back
NM_006445.3:c.2013A>G
p.Thr671= · PRPF8
ACMG/AMP
0%
complete
Final classification
VUS
BP4
PRPF8
c.2013A>G
p.Thr671=
synonymous · exon 15

PRPF8 encodes a core component of the spliceosome, the cellular machinery that removes introns from pre-mRNA during gene expression; it is essential for the catalytic second step of pre-mRNA splicing and helps assemble splicing complexes through its WD repeat domains. Mutations in PRPF8 are a known cause of autosomal dominant retinitis pigmentosa, an inherited form of progressive vision loss. In cancer, altered PRPF8 splicing disrupts the proofreading function of the spliceosome and is often found alongside TP53 loss-of-function mutations, implicating it in tumor development as a tumor suppressor.

This variant

PRPF8 encodes a core spliceosome component; its mutations cause autosomal dominant retinitis pigmentosa (RP13), and altered splicing is implicated in cancer alongside TP53 loss. This synonymous variant (p.Thr671=) shows no predicted splice impact (SpliceAI max delta 0.004) and no pathogenic evidence, yet it cannot be classified benign because its borderline South Asian population frequency and lack of functional data leave uncertainty. The VUS classification means neither pathogenic nor benign significance is established for this variant.

Transcript
NM_006445.3
HGVS · transcript:coding
NM_006445.3:c.2013A>G
GRCh38
chr17:1677144 T>C
GRCh37
chr17:1580438 T>C
Variant of Uncertain Significance: the only met criterion is BP4 (supporting, benign direction; SpliceAI max delta 0.004), which alone matches no Benign or Likely Benign combination rule. Flagged for human review: borderline South Asian frequency (BS1) and homozygous gnomAD observations (BS2).
Classification rationale
BP4 VUS
PRPF8 c.2013A>G synonymous · exon 15

BP4 (Supporting): synonymous variant (p.Thr671=) with SpliceAI max delta 0.004, far below the 0.1 threshold, indicating no predicted splice impact. Overall: Variant of Uncertain Significance - a single supporting benign criterion matches no Pathogenic, Likely Pathogenic, Benign, or Likely Benign combination rule.

BP4 → VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_006445.3 · variants mapped to exon structure
PRPF8 NM_006445.3
Fetching transcript structure from UCSC…
Applied criteria · 1 applied · 18 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 1
Strength Supporting Moderate Strong Very strong
BP4 supporting Benign
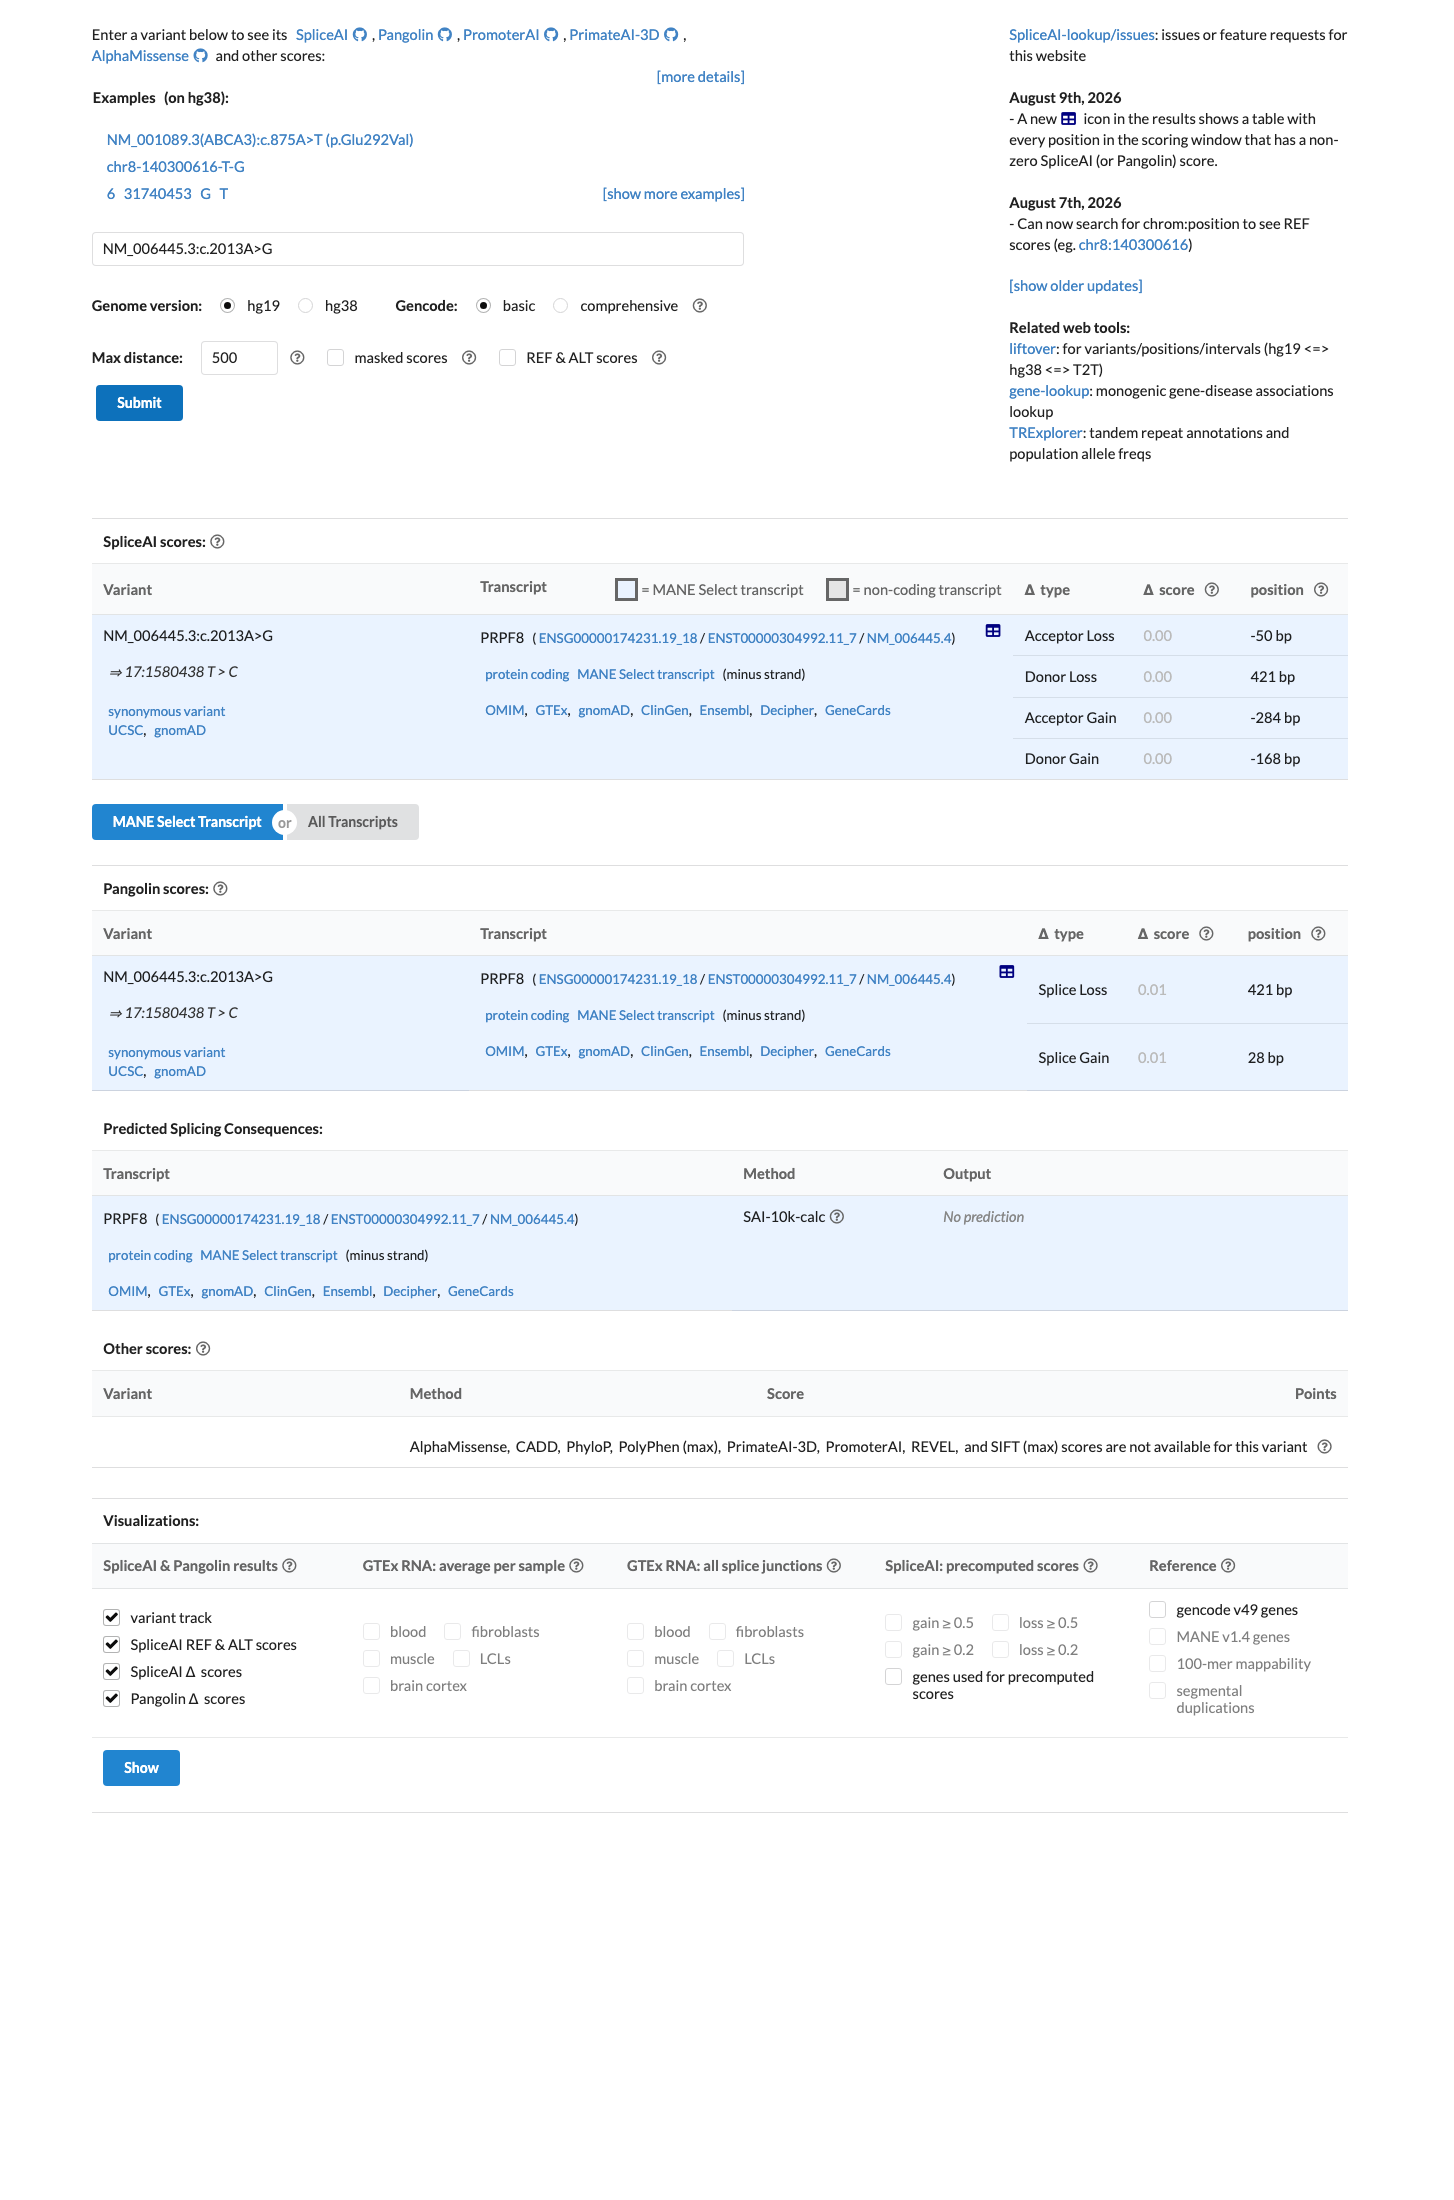
Met (Supporting): SpliceAI max delta 0.004 is far below the <0.1 BP4 threshold, indicating no predicted splice impact.
SpliceAI lookup (spliceai; tool publication Jaganathan et al. 2019, PMID 30661751, Cell 176:535-548) reports max_delta_score 0.004 for NM_006445.3:c.2013A>G.Governing PP3/BP4 calibration (output/generic_acmg_classification_rules.md, source basis Richards et al. 2015 PMID 25741868; SpliceAI thresholds as per SVI recommendations reflected in VCEP CSPECs, e.g. ATM, per Walker et al. 2023): for synonymous/non-missense variants SpliceAI max delta < 0.1 -> BP4 (supporting). Observed 0.004 < 0.1, so BP4 is met (supporting).Per the governing framework, exactly one mechanism is evaluated per variant and a synonymous variant is scored with SpliceAI only; the missense path (REVEL) is not applicable, and no REVEL/BayesDel/aGVGD scores are available for this variant (REVEL and BayesDel lookups found=false).
Assessed · not applied · 8 not met · 10 not assessed
Pathogenic
PS2 Not assessed: no de novo observation with parental testing exists for this variant in any clinical source.
PS3 Not assessed: no functional assay data exist; SpliceAI (max delta 0.004) is in-silico and cannot substitute.
PS4 Not assessed: no case-control or affected-cohort frequency exists; gnomAD documents only general-population frequency.
PM2 Not met: highest subpopulation frequency (South Asian 0.284-0.320%) far exceeds the 0.1% threshold, so the variant is not rare.
PM6 Not assessed: no assumed de novo event or parental testing data exist for this variant.
PP1 Not assessed: no segregation analysis or affected-family genotype data exist for this variant.
PP3 Not met: SpliceAI max delta 0.004 is far below the >0.2 PP3 threshold; no other computational evidence supports pathogenicity.
PP4 Not assessed: no clinical case or phenotype data exist to evaluate phenotype specificity for this variant.
PP5 Not met: no expert-panel ClinVar pathogenic classification exists; the sole submission is a single-laboratory Benign label.
Benign
BA1 Not met: maximum allele frequency (South Asian 0.32%) is far below the >1% BA1 threshold.
BS1 Not met: population-maximum allele frequencies (grpmax FAF 0.256-0.269%) stay below the 0.3% threshold.
BS2 Not met: gnomAD homozygotes (4 in v4.1) do not satisfy the healthy-adult phenotype requirement.
BS3 Not assessed: no well-established functional studies exist; SpliceAI max delta 0.004 is in-silico, not a validated assay.
BS4 Not assessed: no affected-family non-segregation data exist for this variant.
BP2 Not assessed: no phase information (trans or cis with a pathogenic variant) is documented for this variant.
BP5 Not assessed: no case-level data exist to evaluate an alternate molecular basis for disease.
BP6 Not met: the Benign ClinVar label comes from a single laboratory, not an expert panel, so it cannot trigger BP6.
BP7 Not met: crediting BP7 would double-count the SpliceAI prediction already used for BP4, and conservation data are absent.
N/A · 9 PVS1 · PS1 · PM1 · PM3 · PM4 · PM5 · PP2 · BP1 · BP3
Research & evidence
Population frequency
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
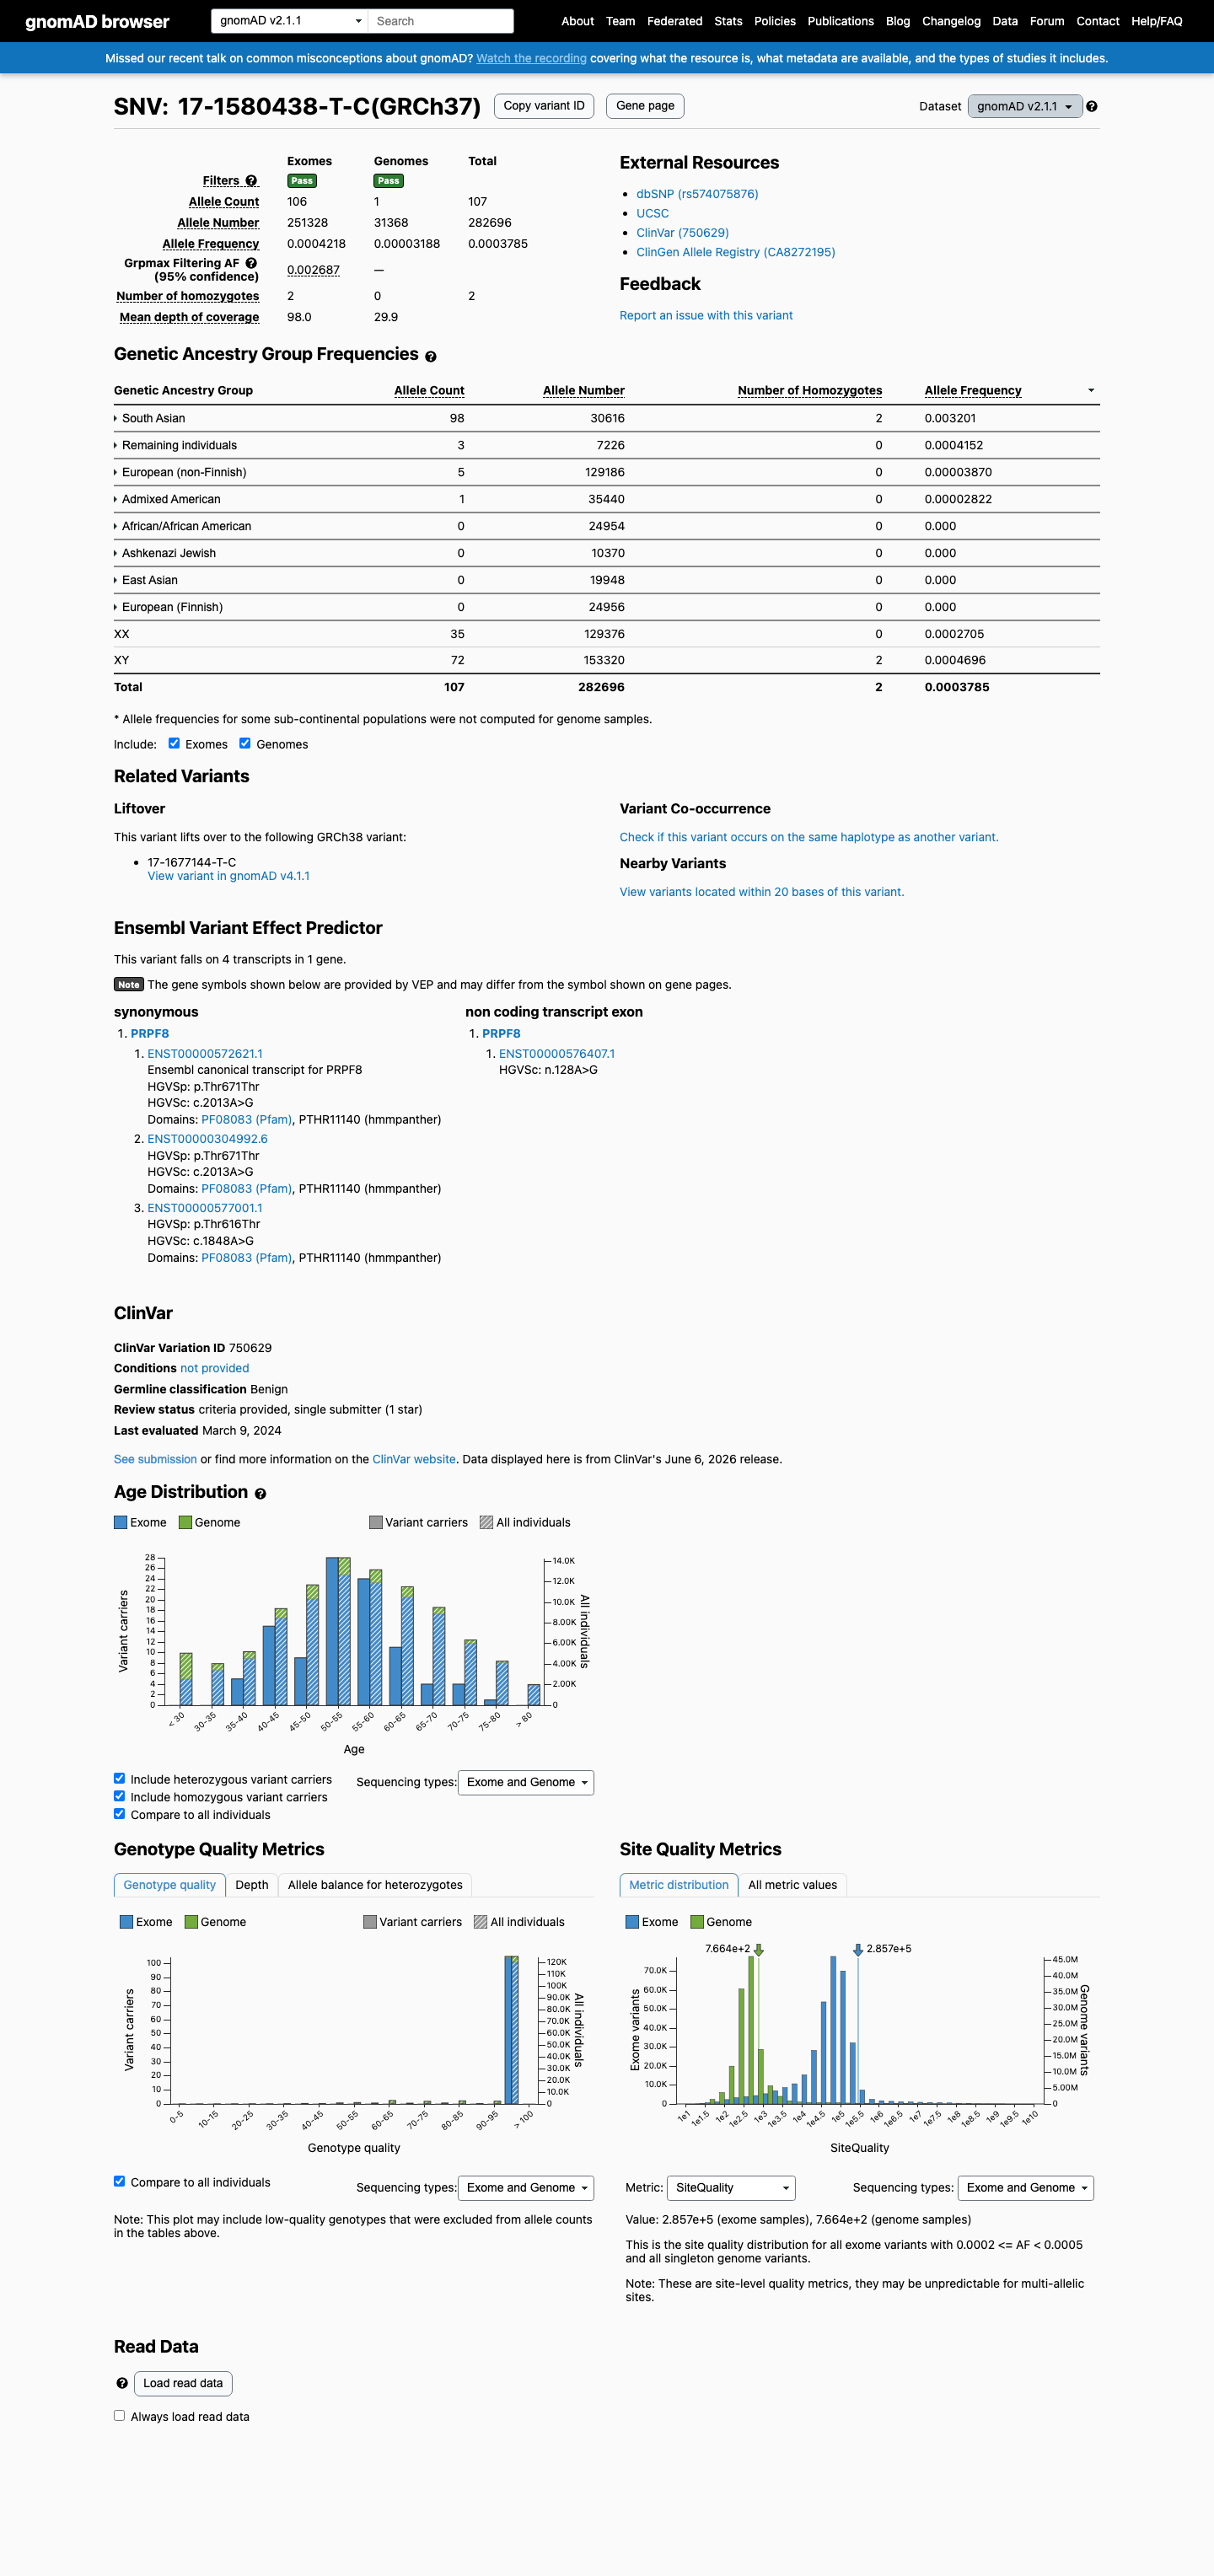
gnomAD v2.1
v4.1
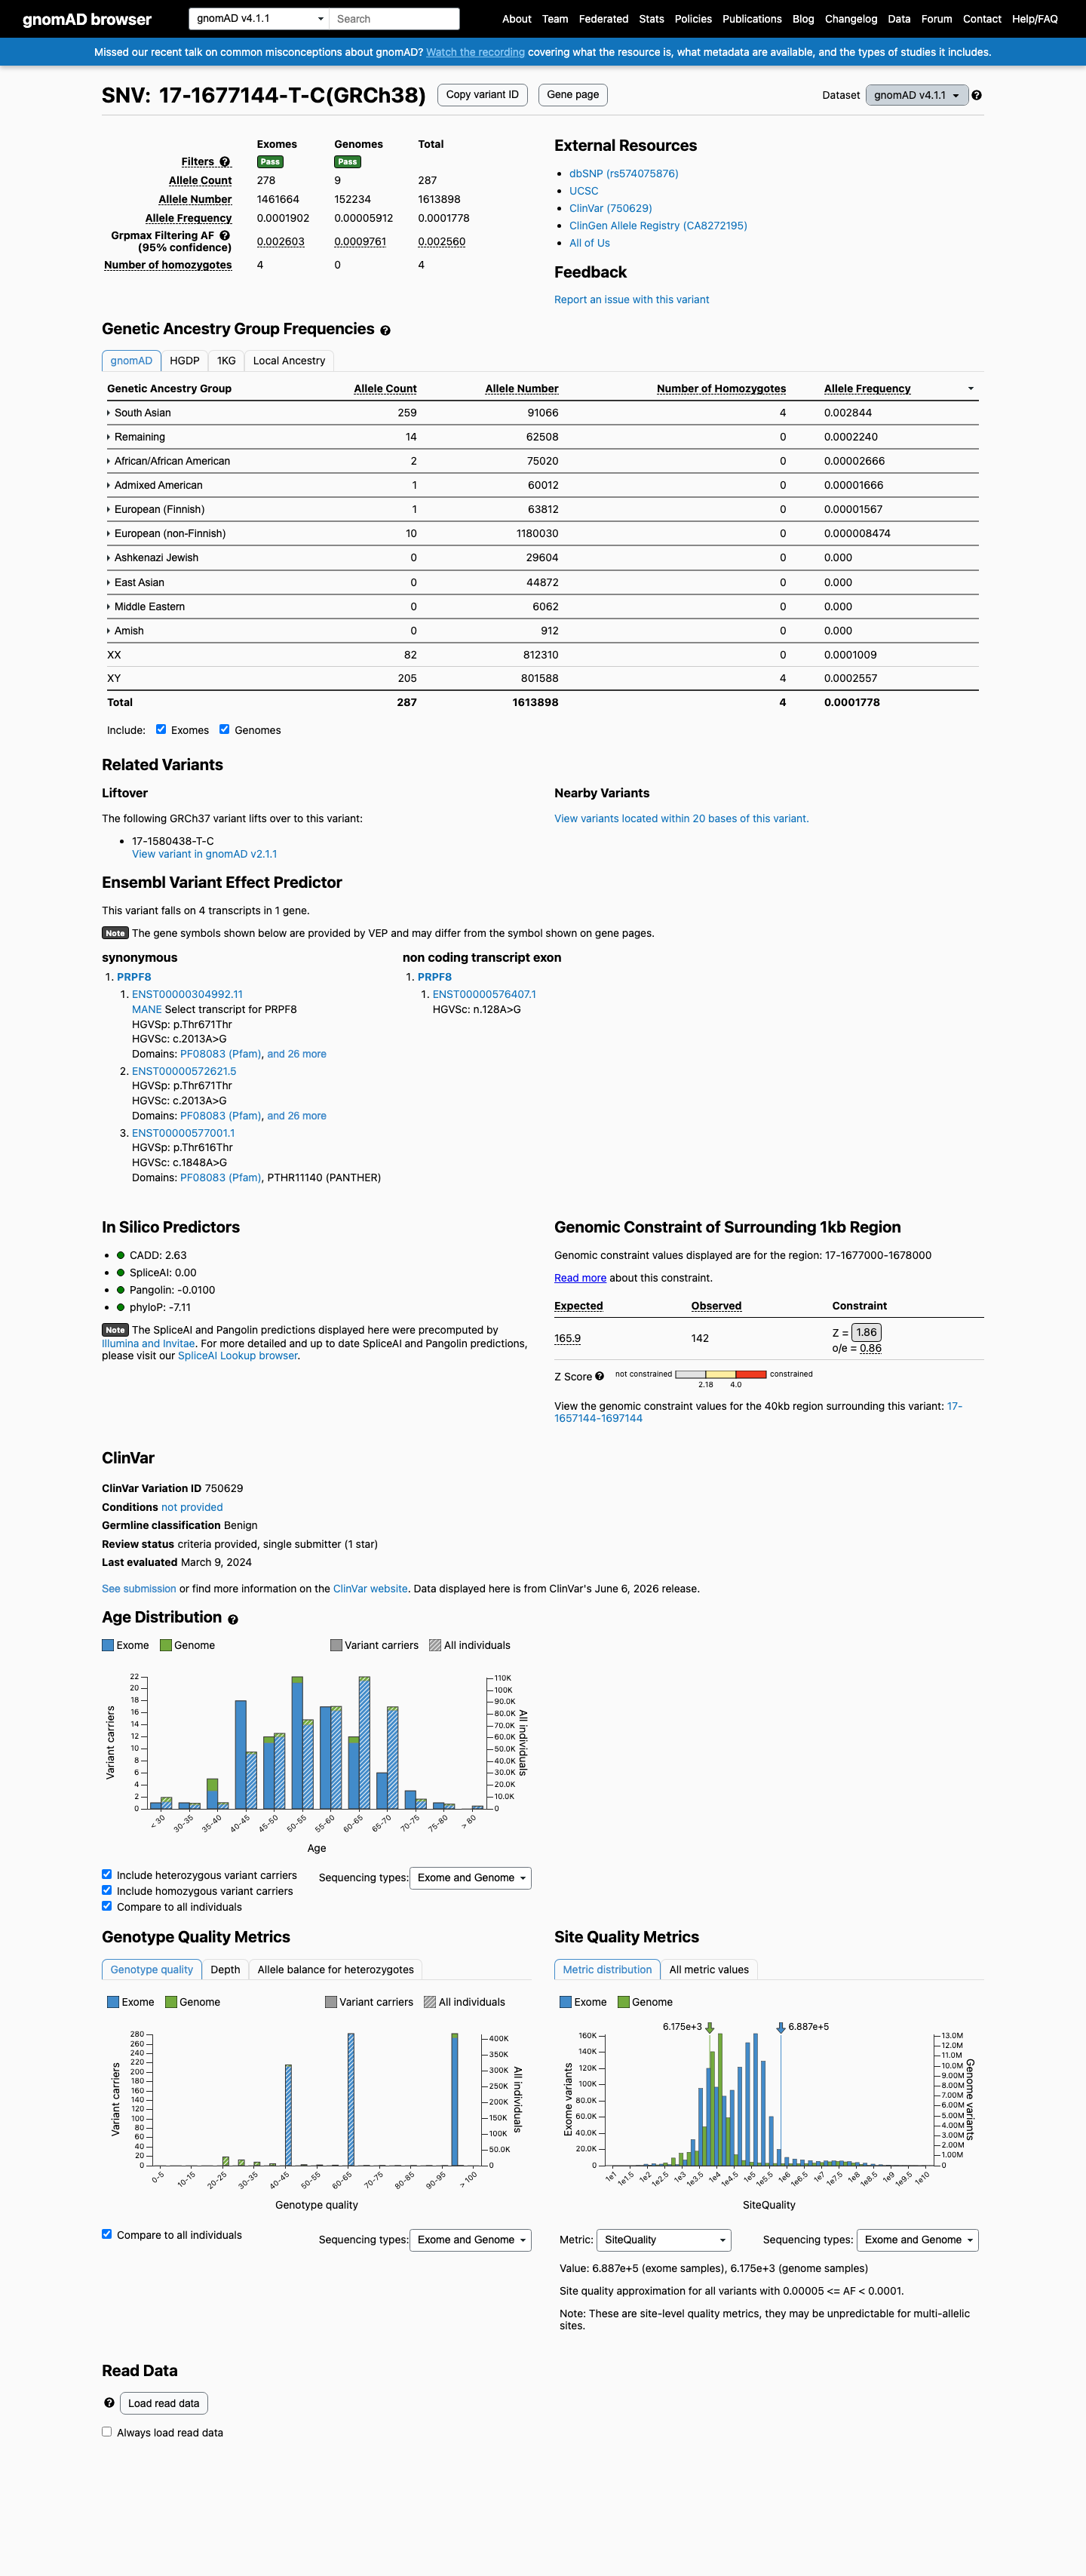
This variant is present in gnomAD v4.1 (AF= 0.00017783; MAF= 0.01778%, 287/1613898 alleles, homozygotes = 4) and has highest observed frequency in the South Asian population (AF= 0.00284409; MAF= 0.28441%, 259/91066 alleles, homozygotes = 4); grpmax FAF= 0.00255955.
v2.1
This variant is present in gnomAD v2.1 (AF= 0.000378498; MAF= 0.03785%, 107/282696 alleles, homozygotes = 2) and has highest observed frequency in the South Asian population (AF= 0.00320094; MAF= 0.32009%, 98/30616 alleles, homozygotes = 2); grpmax FAF= 0.00268731.
🇨🇦 CA
This variant is present in gnomAD-Canada v1.0 (AF= 0.0001085894233901618, 2/18418 alleles, homozygotes = 0).
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.018% · 287 / 1,613,898
4 hom · FAF 0.26%
South Asian
259 / 91,066
0.28%
4 hom
Remaining individuals
14 / 62,508
0.022%
African/African American
2 / 75,020
0.0027%
Admixed American
1 / 60,012
0.0017%
European (Finnish)
1 / 63,812
0.0016%
European (non-Finnish)
10 / 1,180,030
0.00085%
+ 4 not observed (Amish, East Asian, Middle Eastern, Ashkenazi Jewish)
gnomAD v2.1
0.038% · 107 / 282,696
2 hom · FAF 0.27%
South Asian
98 / 30,616
0.32%
2 hom
Remaining individuals
3 / 7,226
0.042%
European (non-Finnish)
5 / 129,186
0.0039%
Admixed American
1 / 35,440
0.0028%
+ 4 not observed (African/African American, Ashkenazi Jewish, East Asian, European (Finnish))
gnomAD Canada 🇨🇦
0.011% · 2 / 18,418
0 hom · FAF 0.026%
South Asian
2 / 1,362
0.15%
+ 8 not observed (African/African American, Latino/Admixed American, Ashkenazi Jewish, East Asian, European (Finnish), Middle Eastern, European (non-Finnish), Remaining individuals)
ClinVar screenshot
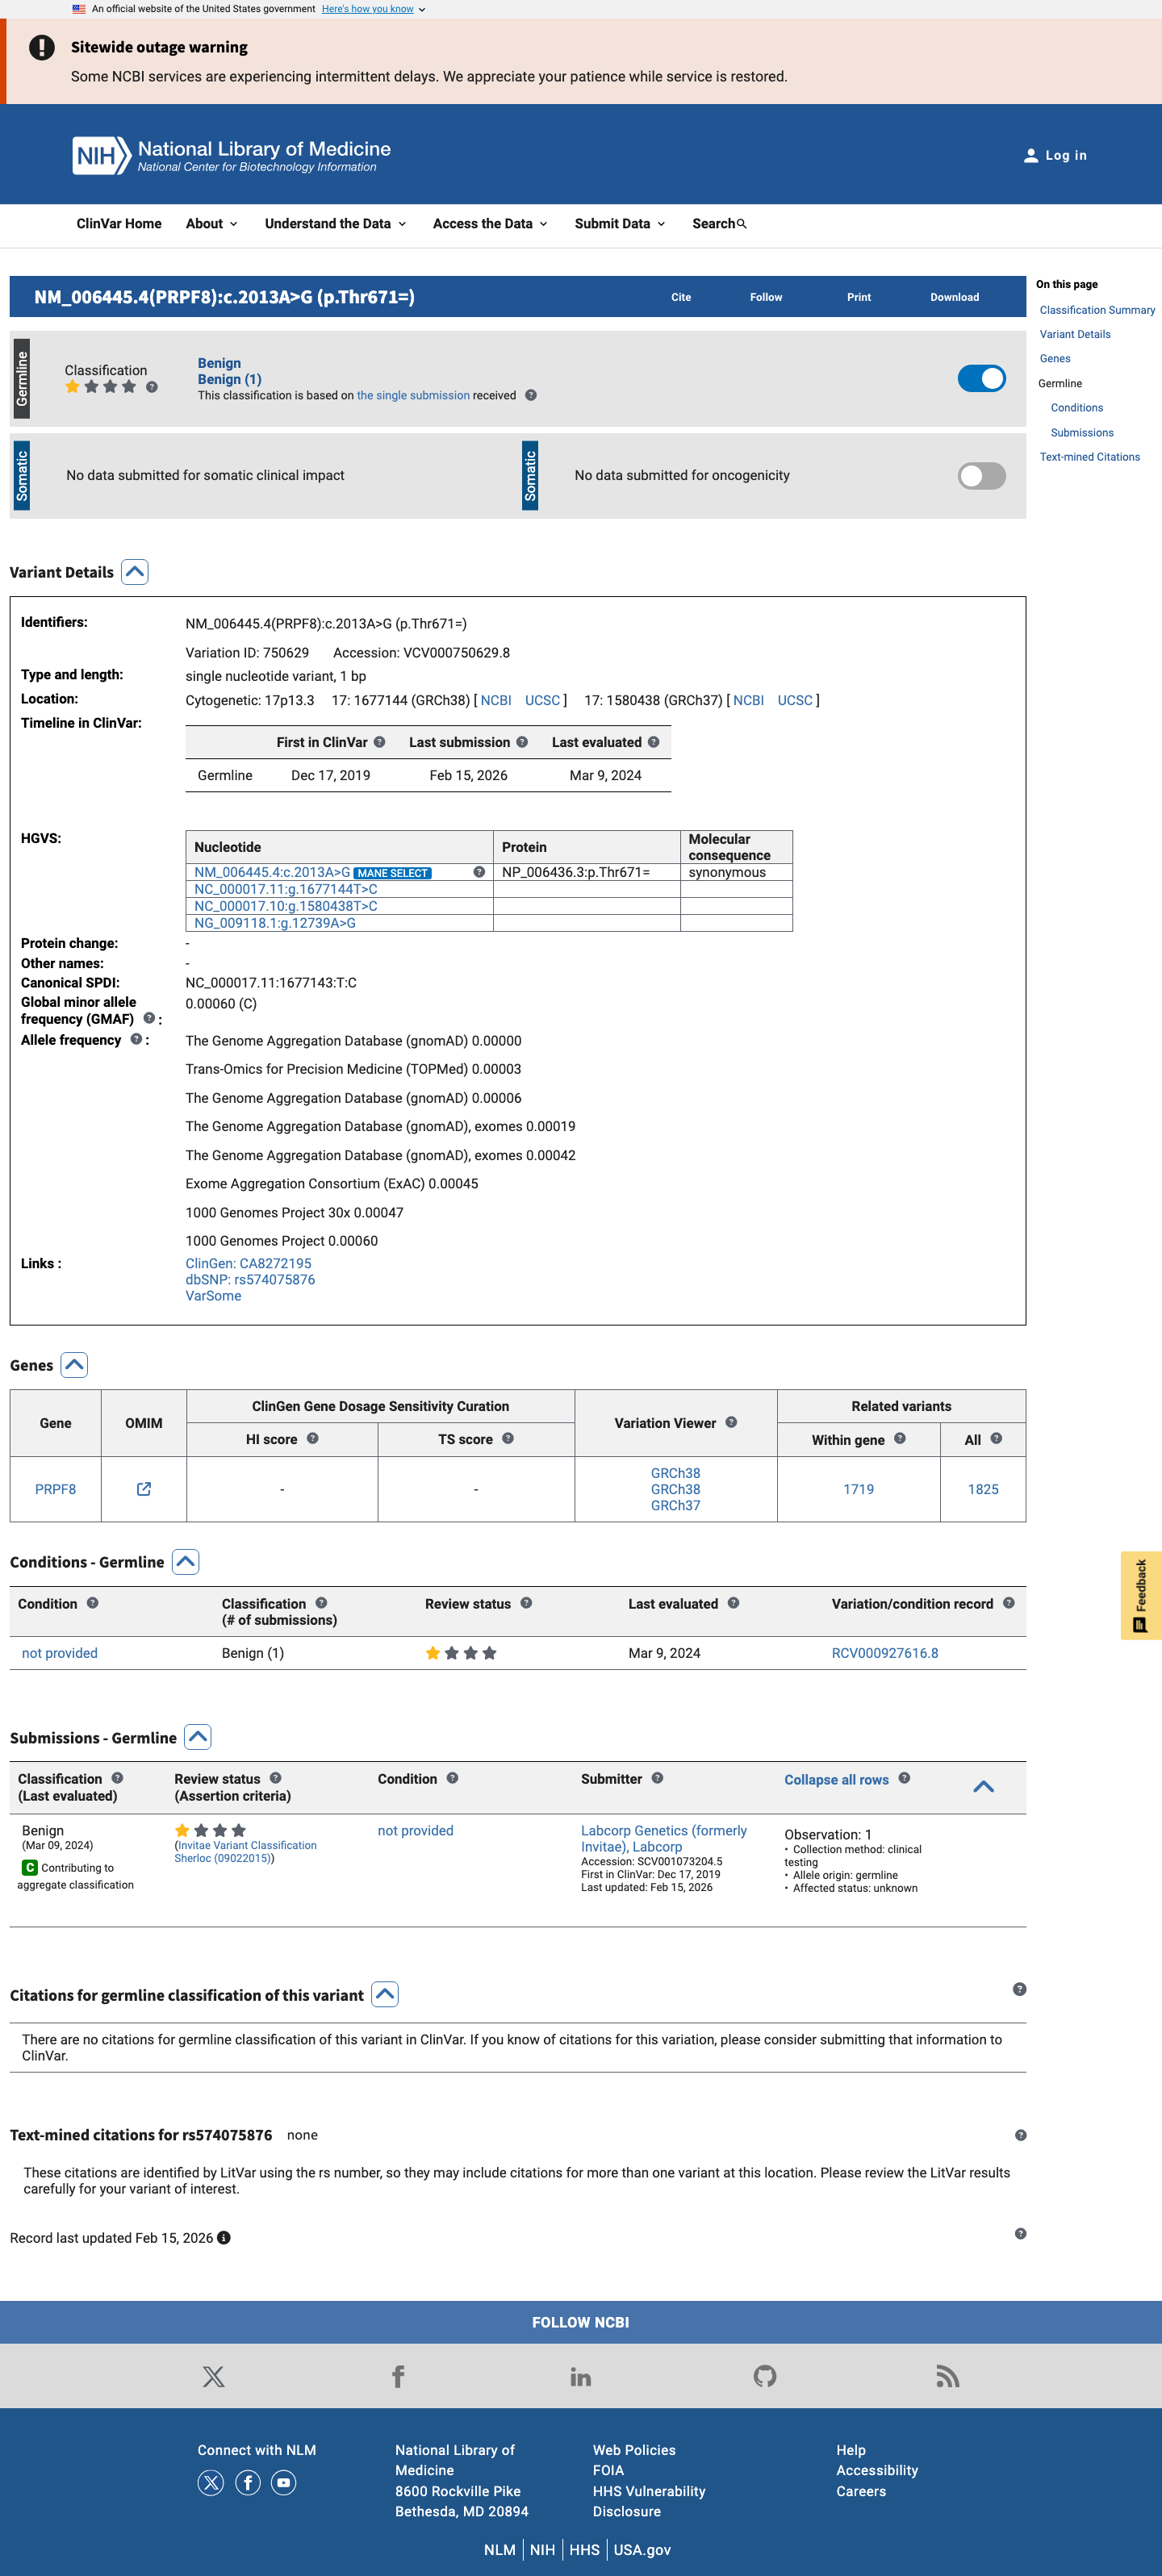
ClinVar
This variant has been reported in ClinVar as Benign (1 clinical laboratory). (ClinVarID = 750629)
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.00).
Functional / OncoKB screenshot
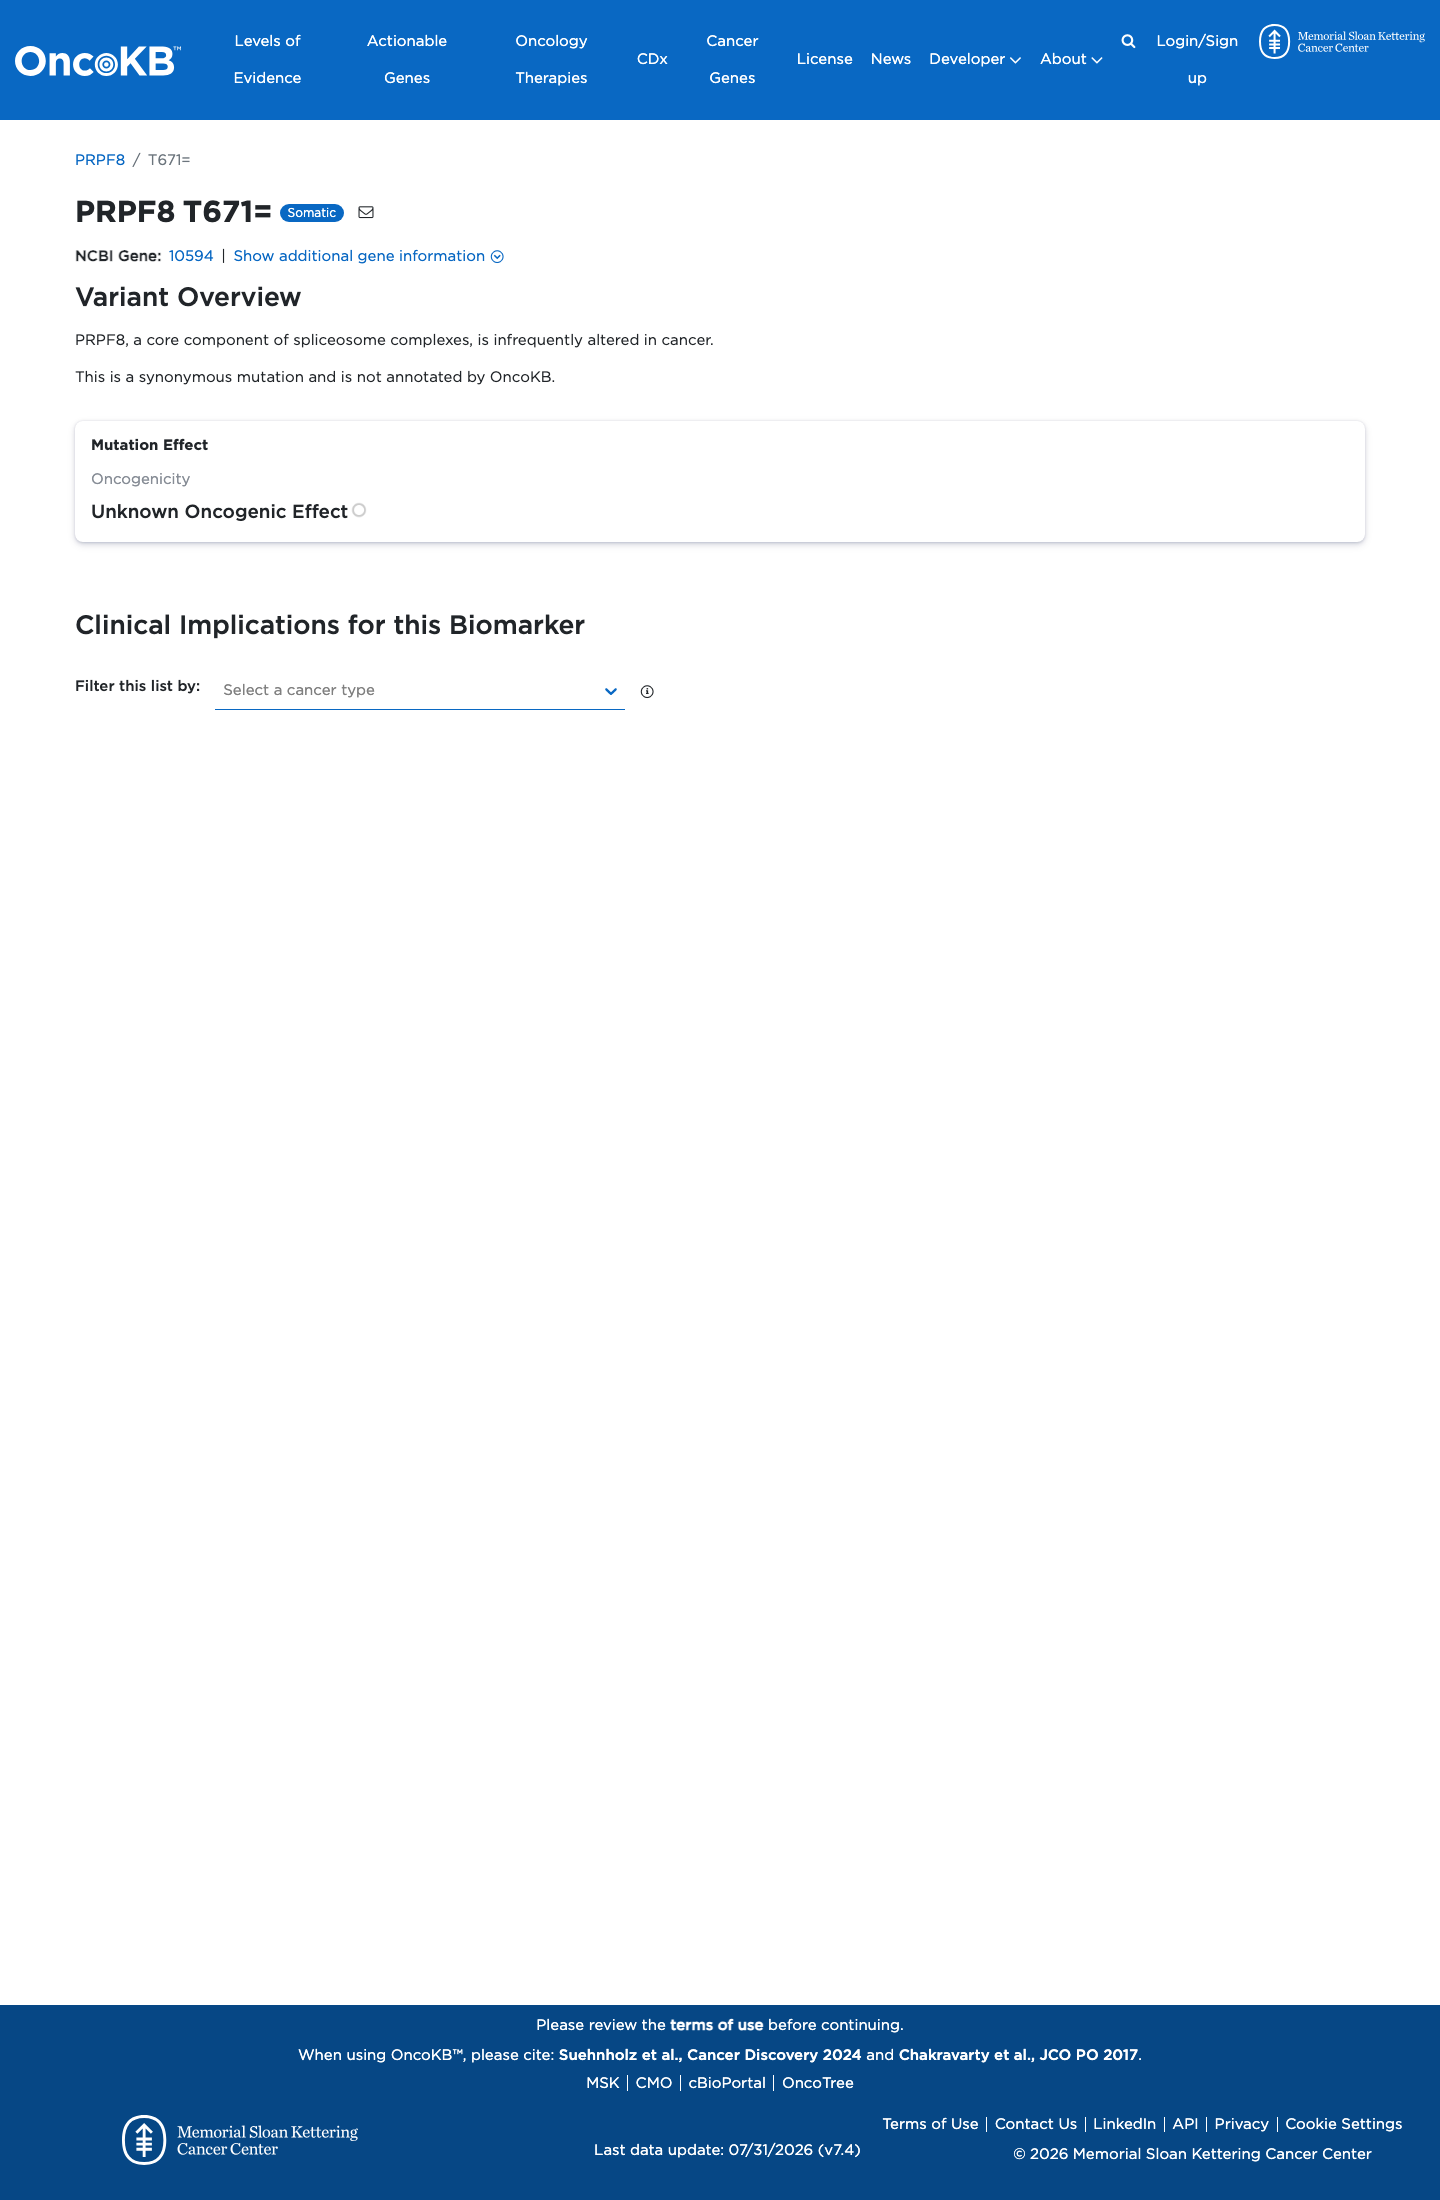
Functional Unknown Oncogenic Effect
OncoKB identified curated literature and non-variant-specific oncogenicity context for review; listed oncogenicity label: Unknown Oncogenic Effect.
OncoKB ↗
COSMIC screenshot
COSMIC
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
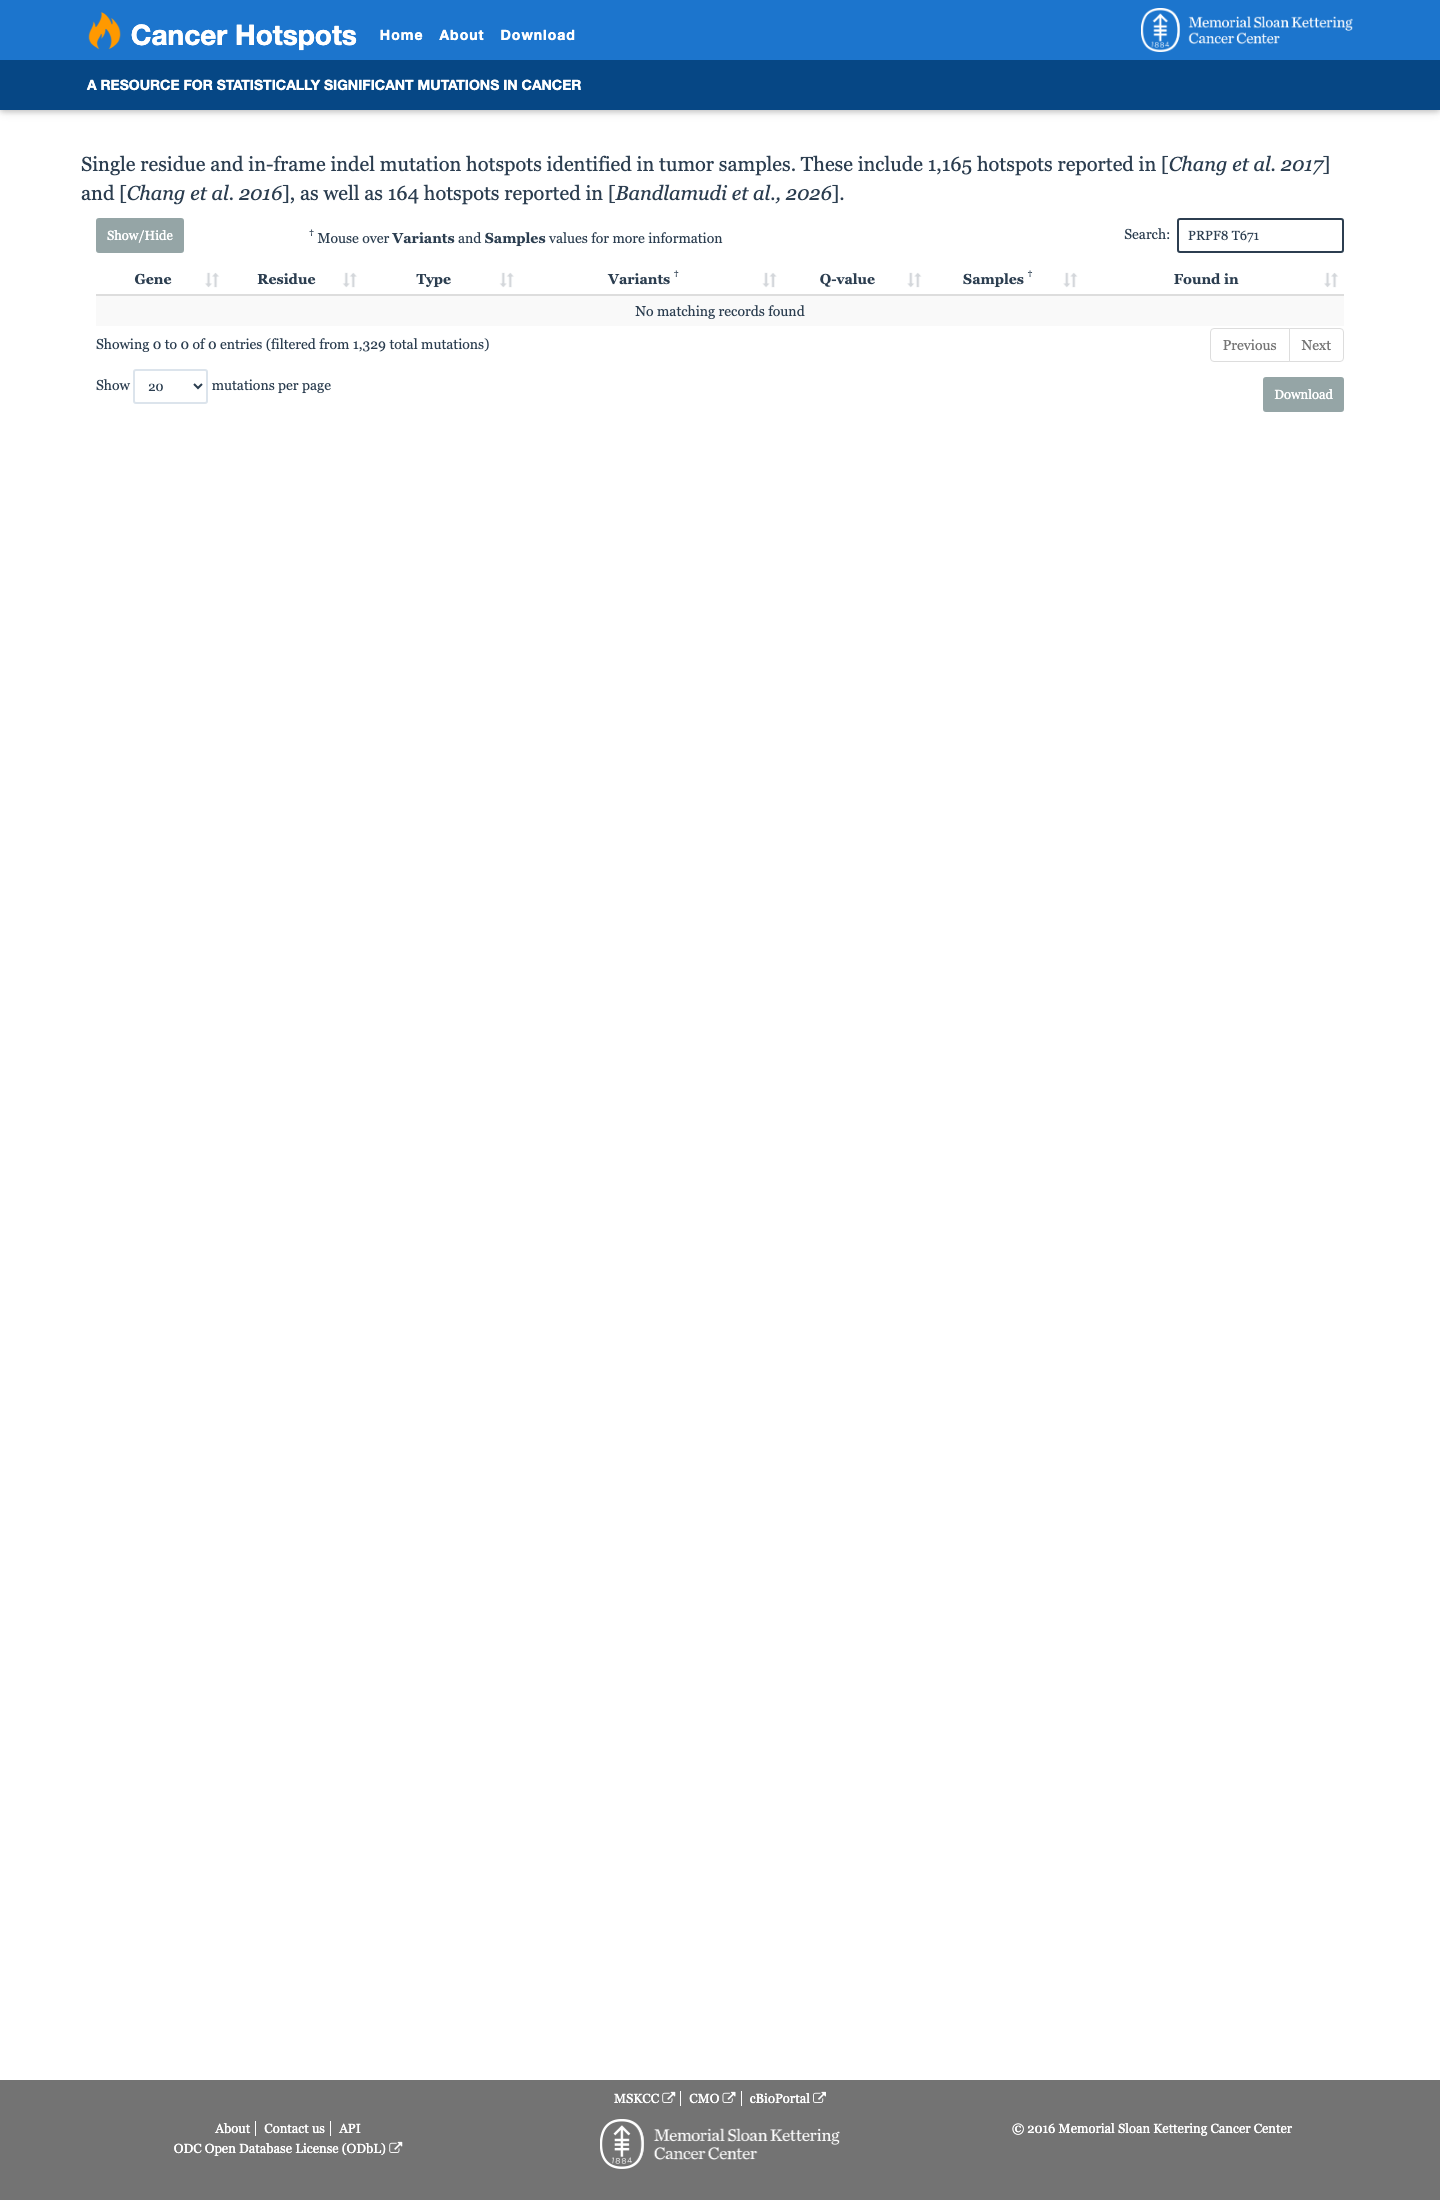
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 1 PMID not cited in assessment
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification criteria. CLINVAR