Back
NM_007294.4:c.3270A>G
p.Gln1090= · BRCA1
0%
complete
Final classification
Likely Benign
BP1BP6
BRCA1
c.3270A>G
p.Gln1090=
synonymous · exon 10

BRCA1 encodes a large nuclear protein that helps maintain genomic stability by coordinating the repair of DNA double-strand breaks through homologous recombination, and it also regulates transcription and the cell cycle. It acts as a tumor suppressor, working with proteins such as RAD51, BRCA2, BARD1, and PALB2 to preserve genome integrity. Inherited alterations in BRCA1 cause hereditary breast and ovarian cancer syndrome, with elevated risks of breast, ovarian, prostate, and pancreatic cancers, and biallelic loss underlies a rare form of Fanconi anemia. Because BRCA1-driven tumors rely on impaired DNA repair, they are treatable with PARP inhibitors.

This variant

BRCA1 encodes a tumor-suppressor protein that coordinates homologous-recombination repair and maintains genomic stability, with inherited pathogenic alterations causing hereditary breast and ovarian cancer syndrome.

Transcript
NM_007294.4
HGVS · transcript:coding
NM_007294.4:c.3270A>G
GRCh38
chr17:43092261 T>C
GRCh37
chr17:41244278 T>C
Likely Benign: BP1 Strong plus BP6 Supporting benign satisfy the ENIGMA BRCA1 Table 3 combination rule.
Classification rationale
BP1BP6 Likely Benign
BRCA1 c.3270A>G synonymous · exon 10

Likely Benign: BP1 Strong is supported by synonymous Q1090 being outside ENIGMA functional domains with SpliceAI maximum delta 0.002. Likely Benign: BP6 Supporting is supported by the exact-variant ClinVar expert-panel Likely benign classification.

BP1 + BP6 Likely Benign
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_007294.4 · variants mapped to exon structure
BRCA1 NM_007294.4
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 14 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
BP1 strong review Benign
Met, strong: synonymous Q1090 is outside ENIGMA BRCA1 domains and SpliceAI max delta 0.002 is below the <=0.1 BP1 threshold.
ENIGMA BP1_Strong rule: apply to silent substitutions outside a potentially clinically important functional domain with SpliceAI <=0.1.Q1090 is outside the ENIGMA BRCA1 domains RING aa 2-101, coiled-coil aa 1391-1424, and BRCT repeats aa 1650-1857.The case SpliceAI result is max delta 0.002, satisfying the governing <=0.1 threshold.
BP6 supporting Benign
Met, Supporting: exact-variant ClinVar expert-panel classification is Likely benign for variation 187419.
ClinVar variation 187419 exactly matches NM_007294.4:c.3270A>G (p.Gln1090=) and is classified Likely benign with review status reviewed by expert panel.The exact protein synonym p.Q1090= is confirmed by the normalized variant record, establishing identity with p.Gln1090=.ClinVar expert panel classification
Assessed · not applied · 5 not met · 9 not assessed
Pathogenic
PS3 Not assessed: no variant-specific calibrated protein or combined mRNA–protein functional assay result was found for c.3270A>G (p.Gln1090=).
PS4 Not assessed: no exact-variant case-control odds ratio, p-value, or confidence interval is available to evaluate the ENIGMA PS4 thresholds.
PM2 Not met: the variant is present with 2 alleles in both governing non-cancer datasets, rather than absent as required for PM2.
PM3 Not assessed: no Fanconi-anemia phenotype, co-occurring pathogenic BRCA1 variant, or phase information is documented for c.3270A>G.
PP1 Not assessed: no affected-relative segregation data or quantitative LR is available to compare with the PP1 Supporting threshold of 2.08:1.
PP3 Not met: SpliceAI maximum delta 0.002 is below the ENIGMA PP3 threshold of >=0.2 for predicted splicing.
PP4 Not assessed: no exact-variant combined multifactorial likelihood ratio is available for comparison with the ENIGMA PP4 threshold of >=2.08:1.
PP5 Not met: exact-variant ClinVar expert-panel classification is Likely benign, not Pathogenic or Likely pathogenic.
Benign
BA1 Not met: maximum non-cancer filter allele frequency was 4.88e-06, below the ENIGMA BA1 threshold of >0.001.
BS1 Not met: maximum non-cancer filter allele frequency was 4.88e-06, below the ENIGMA BS1 Supporting threshold of >0.00002.
BS2 Not assessed: gnomAD reports zero homozygotes, but ENIGMA excludes population observations and no qualifying phenotyped adult was documented.
BS3 Not assessed: no variant-specific calibrated benign protein or combined mRNA–protein assay result was found for c.3270A>G (p.Gln1090=).
BS4 Not assessed: no non-segregating affected relatives or quantitative LR is available to compare with the BS4 Supporting threshold of 0.48.
BP5 Not assessed: no exact-variant multifactorial likelihood ratio is available to test against the BP5 Supporting threshold of <=0.48:1.
N/A · 12 PVS1 · PS1 · PS2 · PM1 · PM4 · PM5 · PM6 · PP2 · BP2 · BP3 · BP4 · BP7
Research & evidence
Population frequency · supports benign
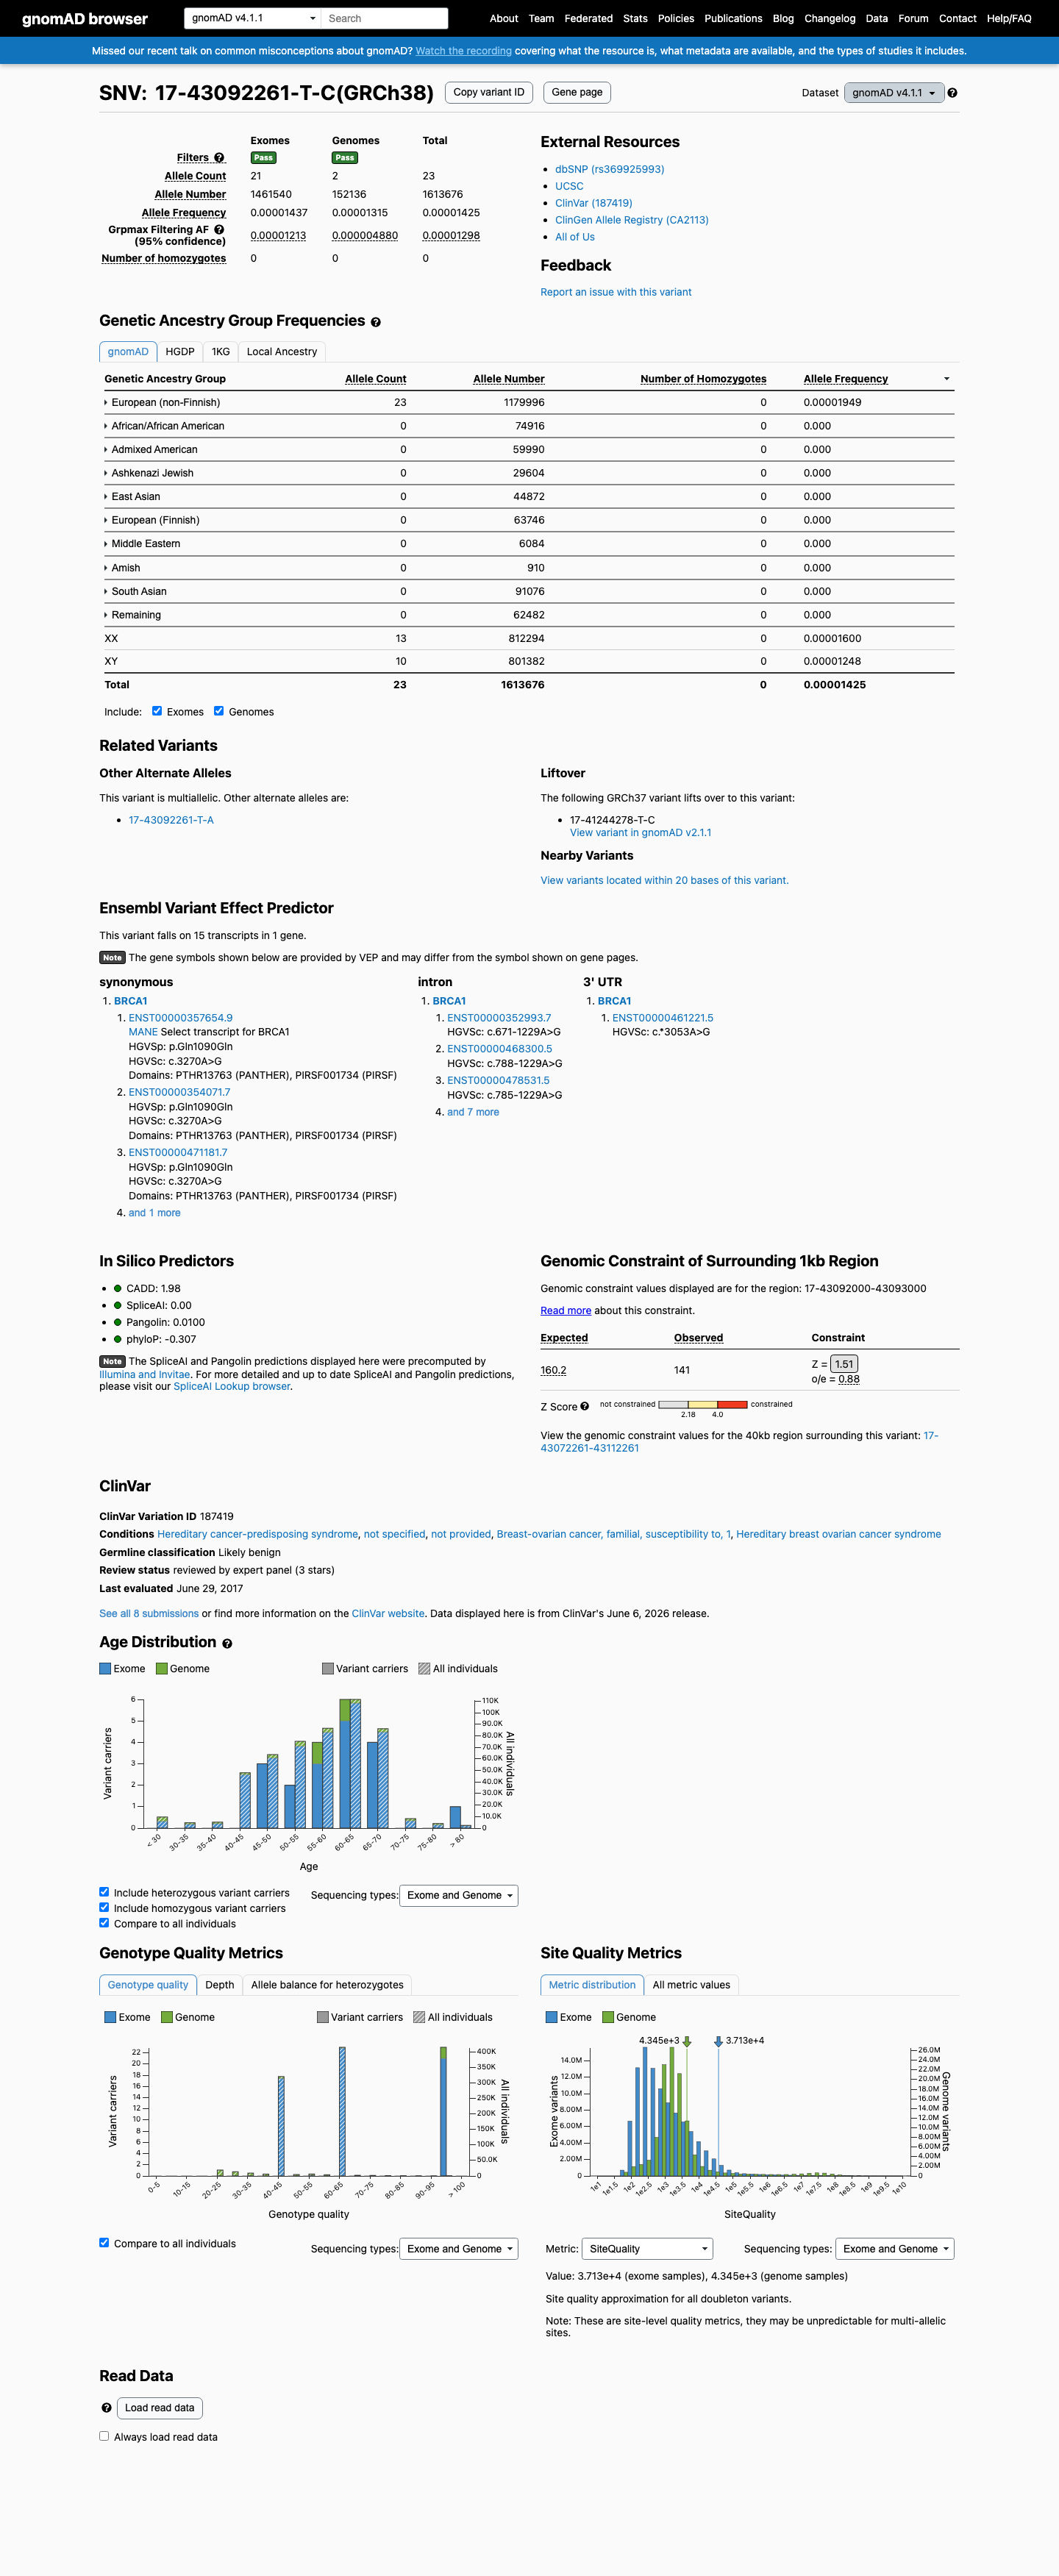
gnomAD v4.1 screenshot
gnomAD v4.1
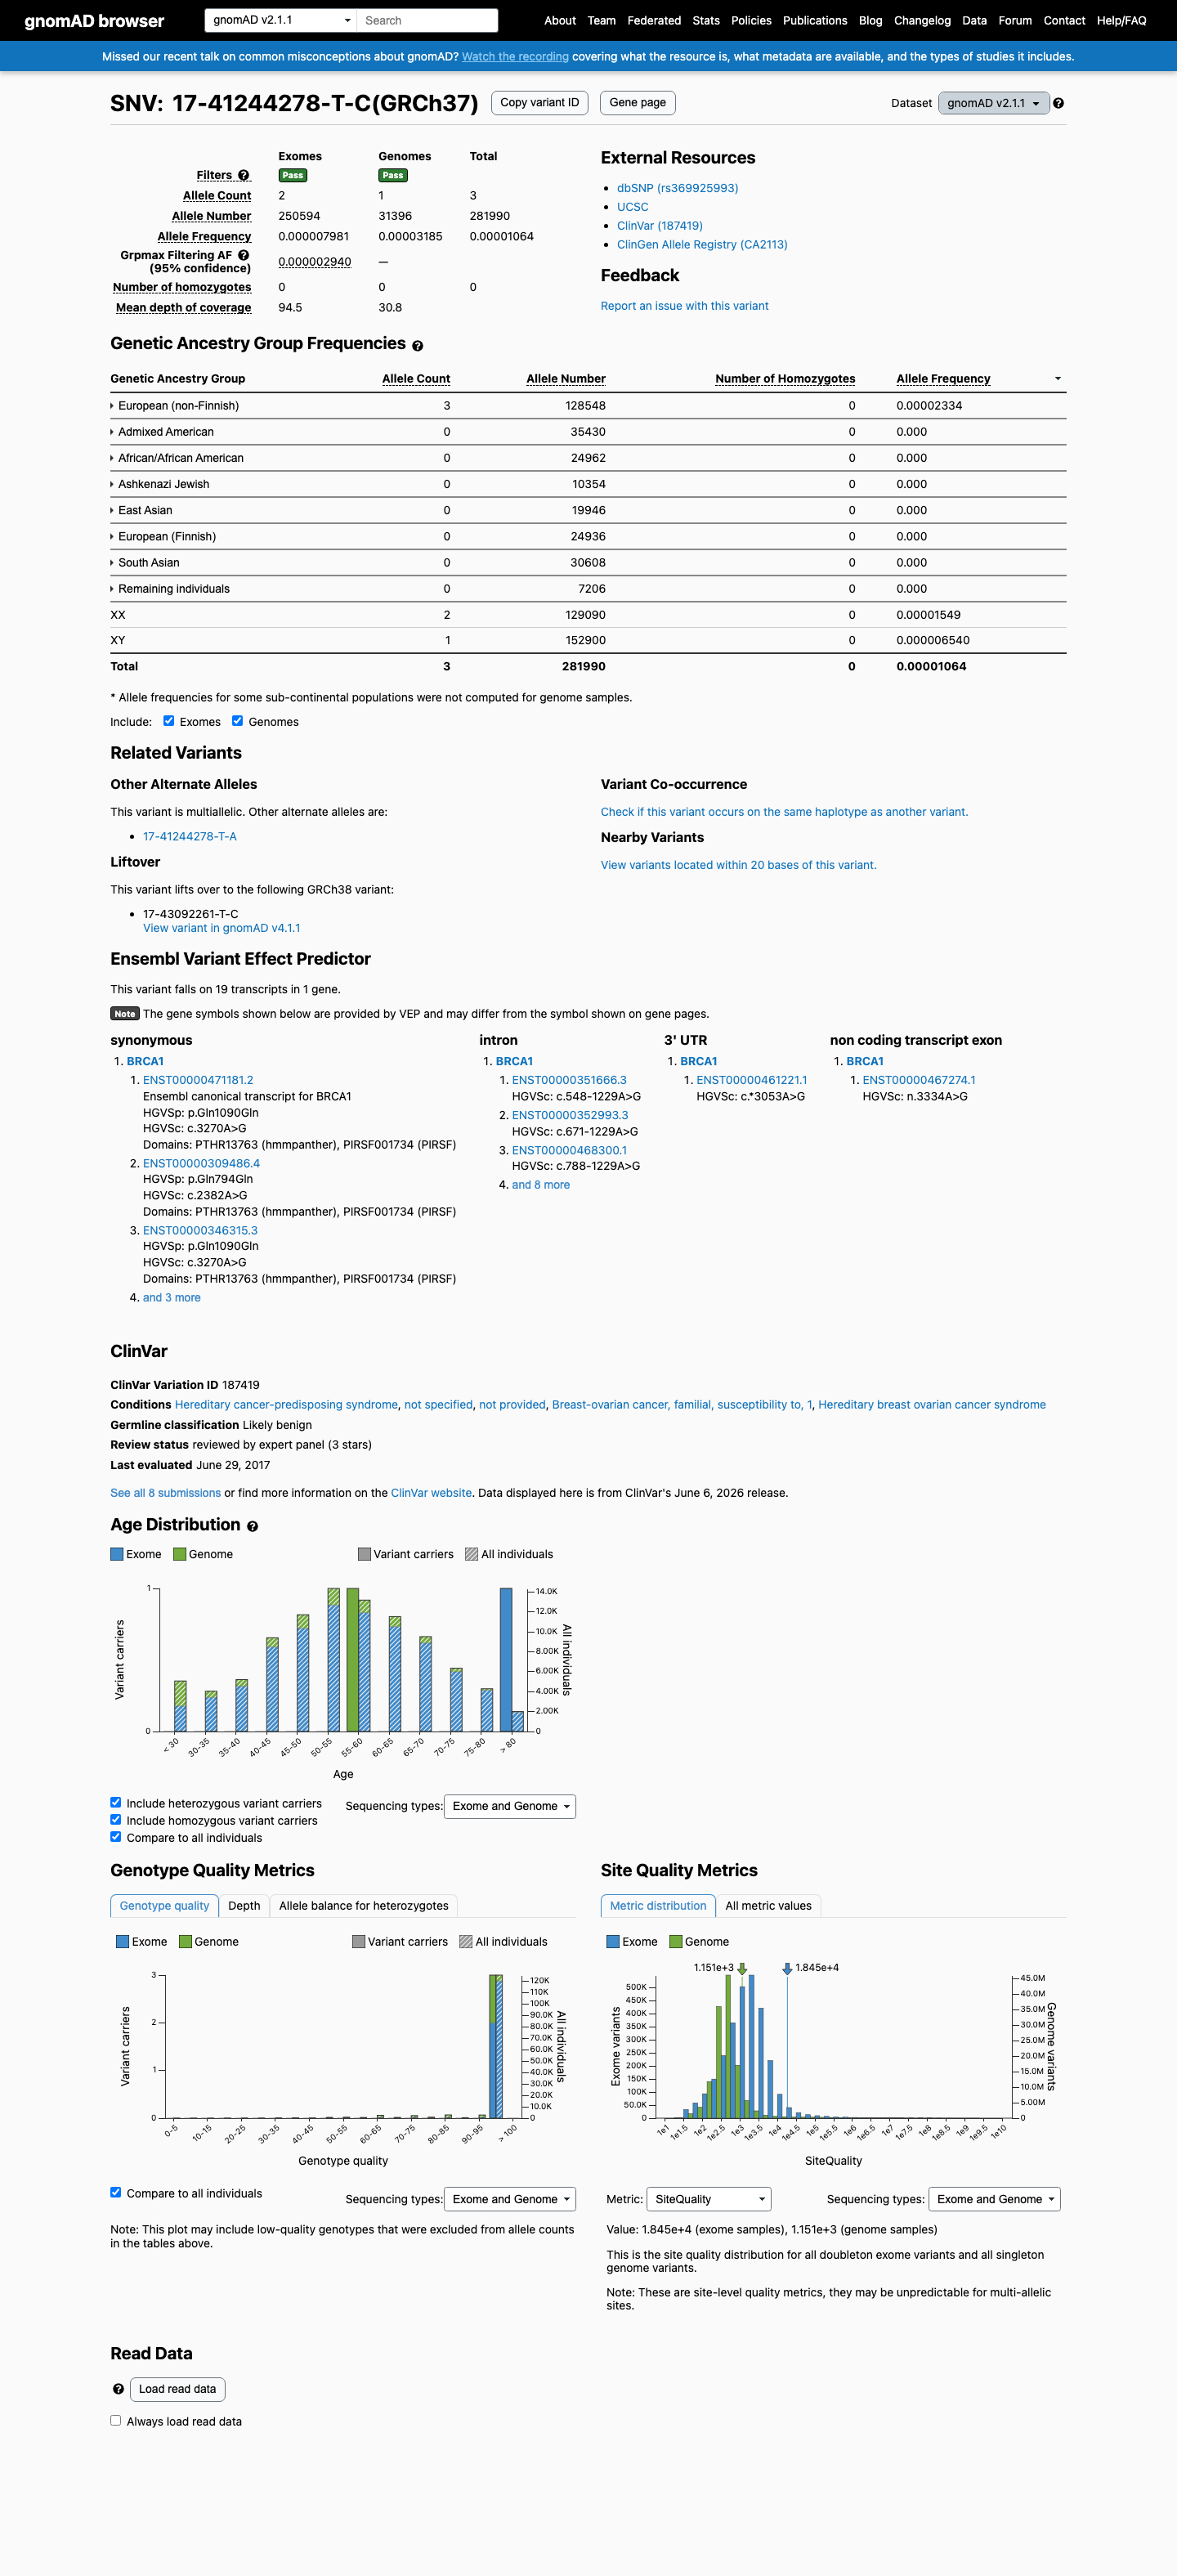
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 1.42532e-05; MAF= 0.00143%, 23/1613676 alleles, homozygotes = 0) and has highest observed frequency in the European (non-Finnish) population (AF= 1.94916e-05; MAF= 0.00195%, 23/1179996 alleles, homozygotes = 0); grpmax FAF= 1.298e-05.
v2.1
This variant is present in gnomAD v2.1 (AF= 1.06387e-05; MAF= 0.00106%, 3/281990 alleles, homozygotes = 0) and has highest observed frequency in the European (non-Finnish) population (AF= 2.33376e-05; MAF= 0.00233%, 3/128548 alleles, homozygotes = 0); grpmax FAF= 2.94e-06.
🇨🇦 CA
Absent from gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.0014% · 23 / 1,613,676
0 hom · FAF 0.0013%
European (non-Finnish)
23 / 1,179,996
0.0019%
+ 9 not observed (Remaining individuals, Admixed American, European (Finnish), Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish, African/African American)
gnomAD v2.1
0.0011% · 3 / 281,990
0 hom · FAF 0.00029%
European (non-Finnish)
3 / 128,548
0.0023%
+ 7 not observed (African/African American, Admixed American, Ashkenazi Jewish, East Asian, European (Finnish), Remaining individuals, South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
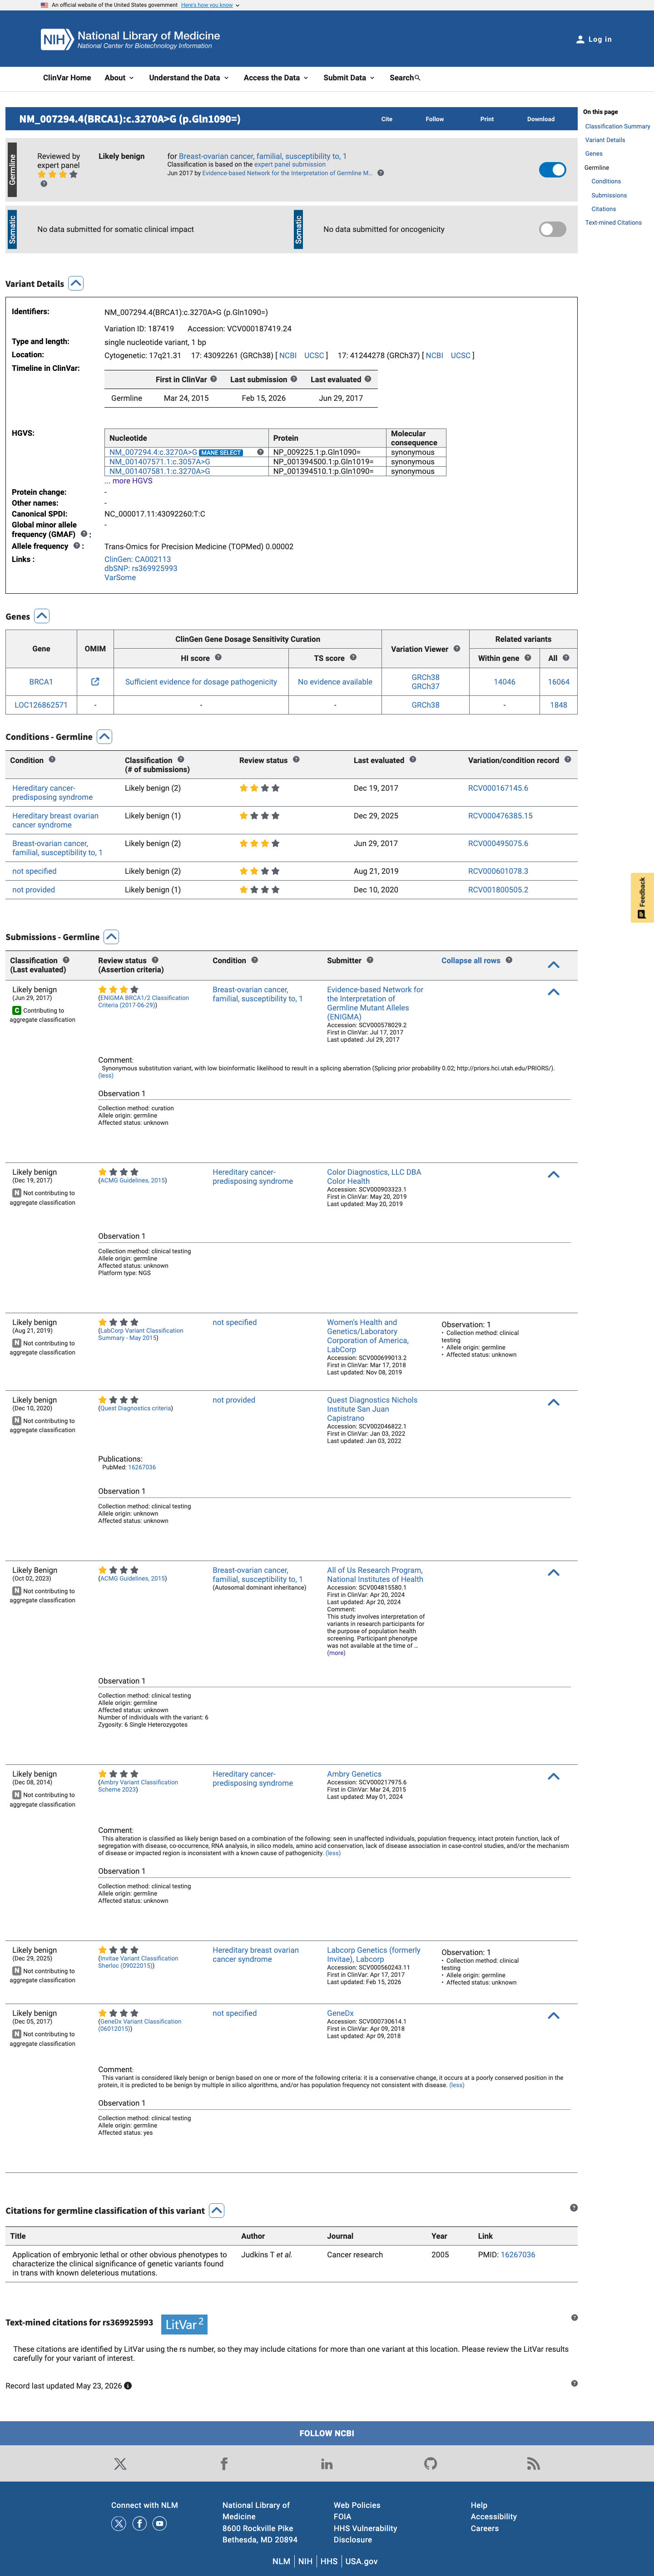
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Likely benign (6 clinical laboratories) and as Likely Benign (1 clinical laboratory) and as Likely benign by Evidence-based Network for the Interpretation of Germline Mutant Alleles (ENIGMA) (expert panel). (ClinVarID = 187419)
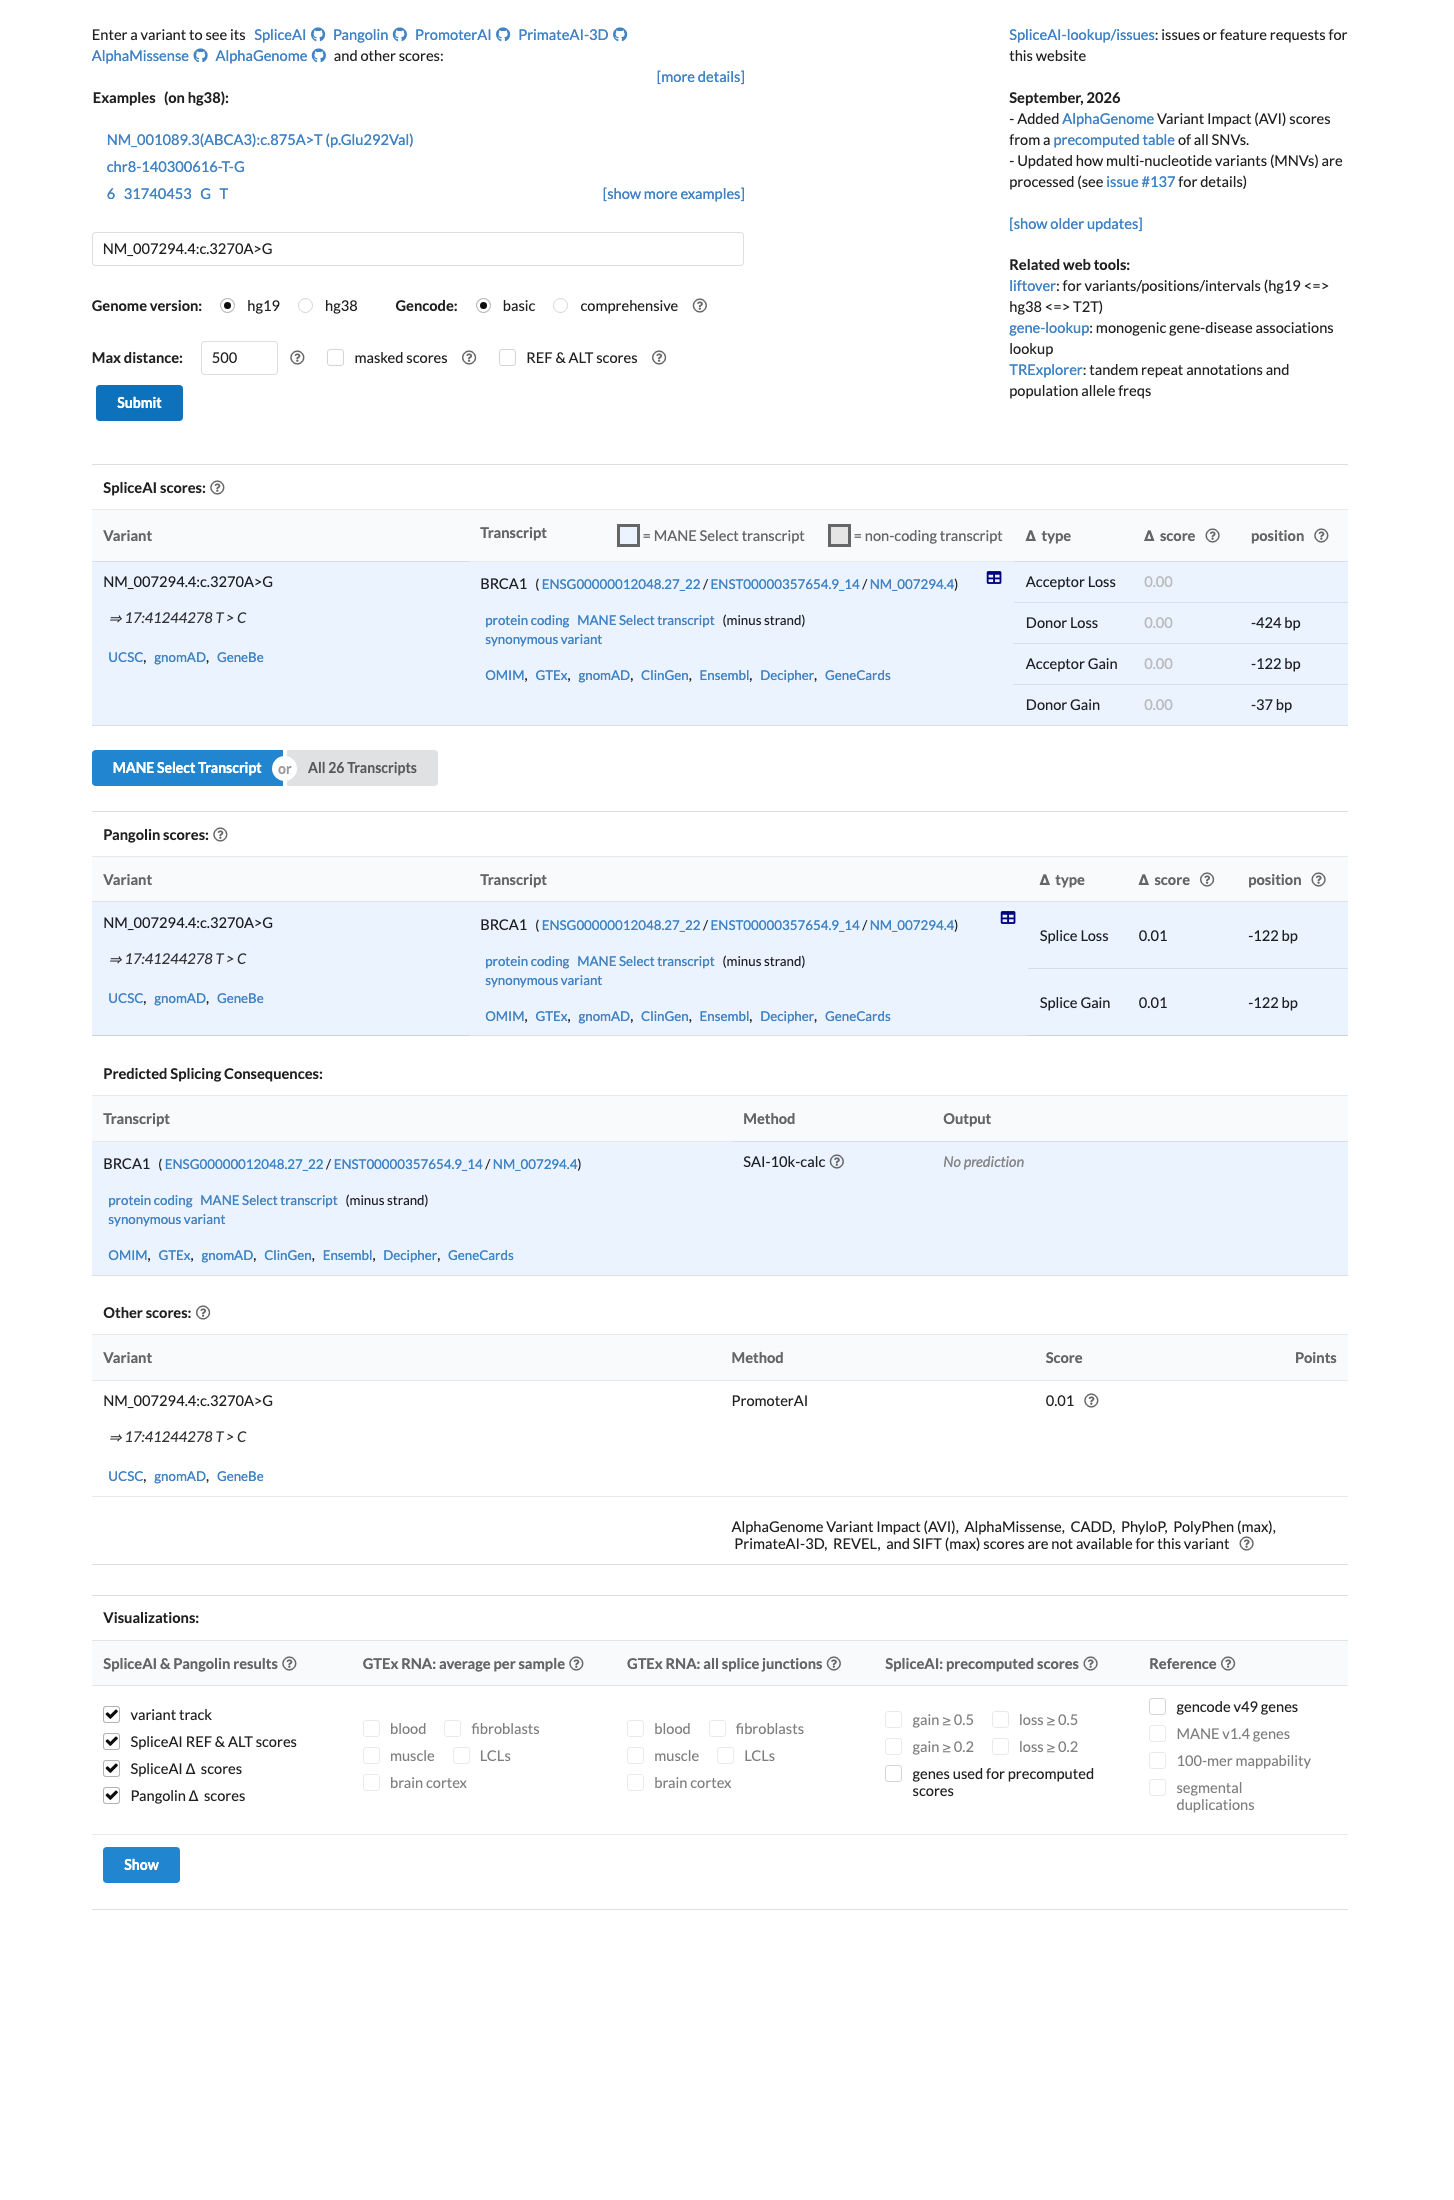
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.00).
Functional / OncoKB screenshot
Functional Unknown Oncogenic Effect
OncoKB identified curated literature and non-variant-specific oncogenicity context for review; listed oncogenicity label: Unknown Oncogenic Effect.
OncoKB ↗
COSMIC screenshot
COSMIC
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
1papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 8 further PMIDs triaged but not cited — see Sources & references.
Application of embryonic lethal or other obvious phenotypes to characterize the clinical significance of genetic variants found in trans with known deleterious mutations.
Searched
c.3270A>GNP_009225.1:p.(Q1090=)p.(Q1090=)Q1090Q
Found
Judkins et al. list the synonymous BRCA1 variant Q1090Q in Table 2 under Haplotype 1 and label it P, defined in the paper as a benign polymorphism; the study reports sequencing data from 55,630 patients.
Variant
✓ Names this variant — characterised directly
Applied to
BP1 strong
Variant-specific benign-polymorphism context is concordant with the benign direction of BP1, while BP1 strength itself is assigned by the ENIGMA domain and SpliceAI rule.
Q1090Q (P) ... The currently reported clinical significance ... of variants are designated as benign polymorphism (P), deleterious mutation (D), or uncertain clinical significance (U).
Location Table 2, Haplotype 1; Table 2 note  ·  Context Clinical BRCA1 whole-gene direct DNA sequencing data from 55,630 patients; variants were assigned to canonical haplotypes using genotype data from 14 haplotype-tagging SNPs.  ·  full text
Sources & reference links
9Sources
CSpec VCEP
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 8 PMIDs not cited in assessment
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology. CLINVAR
26467025 ↗ A Standardized DNA Variant Scoring System for Pathogenicity Assessments in Mendelian Disorders. CLINVAR
12692171 ↗ American Society of Clinical Oncology policy statement update: genetic testing for cancer susceptibility. CLINVAR
20065170 ↗ American Society of Clinical Oncology policy statement update: genetic and genomic testing for cancer susceptibility. CLINVAR
24366376 ↗ Risk assessment, genetic counseling, and genetic testing for BRCA-related cancer in women: U.S. Preventive Services Task Force recommendation statement. CLINVAR
24493721 ↗ American Society of Clinical Oncology Expert Statement: collection and use of a cancer family history for oncology providers. CLINVAR
31479213 ↗ PMID 31479213 CLINVAR
28492532 ↗ Sherloc: a comprehensive refinement of the ACMG-AMP variant classification criteria. CLINVAR