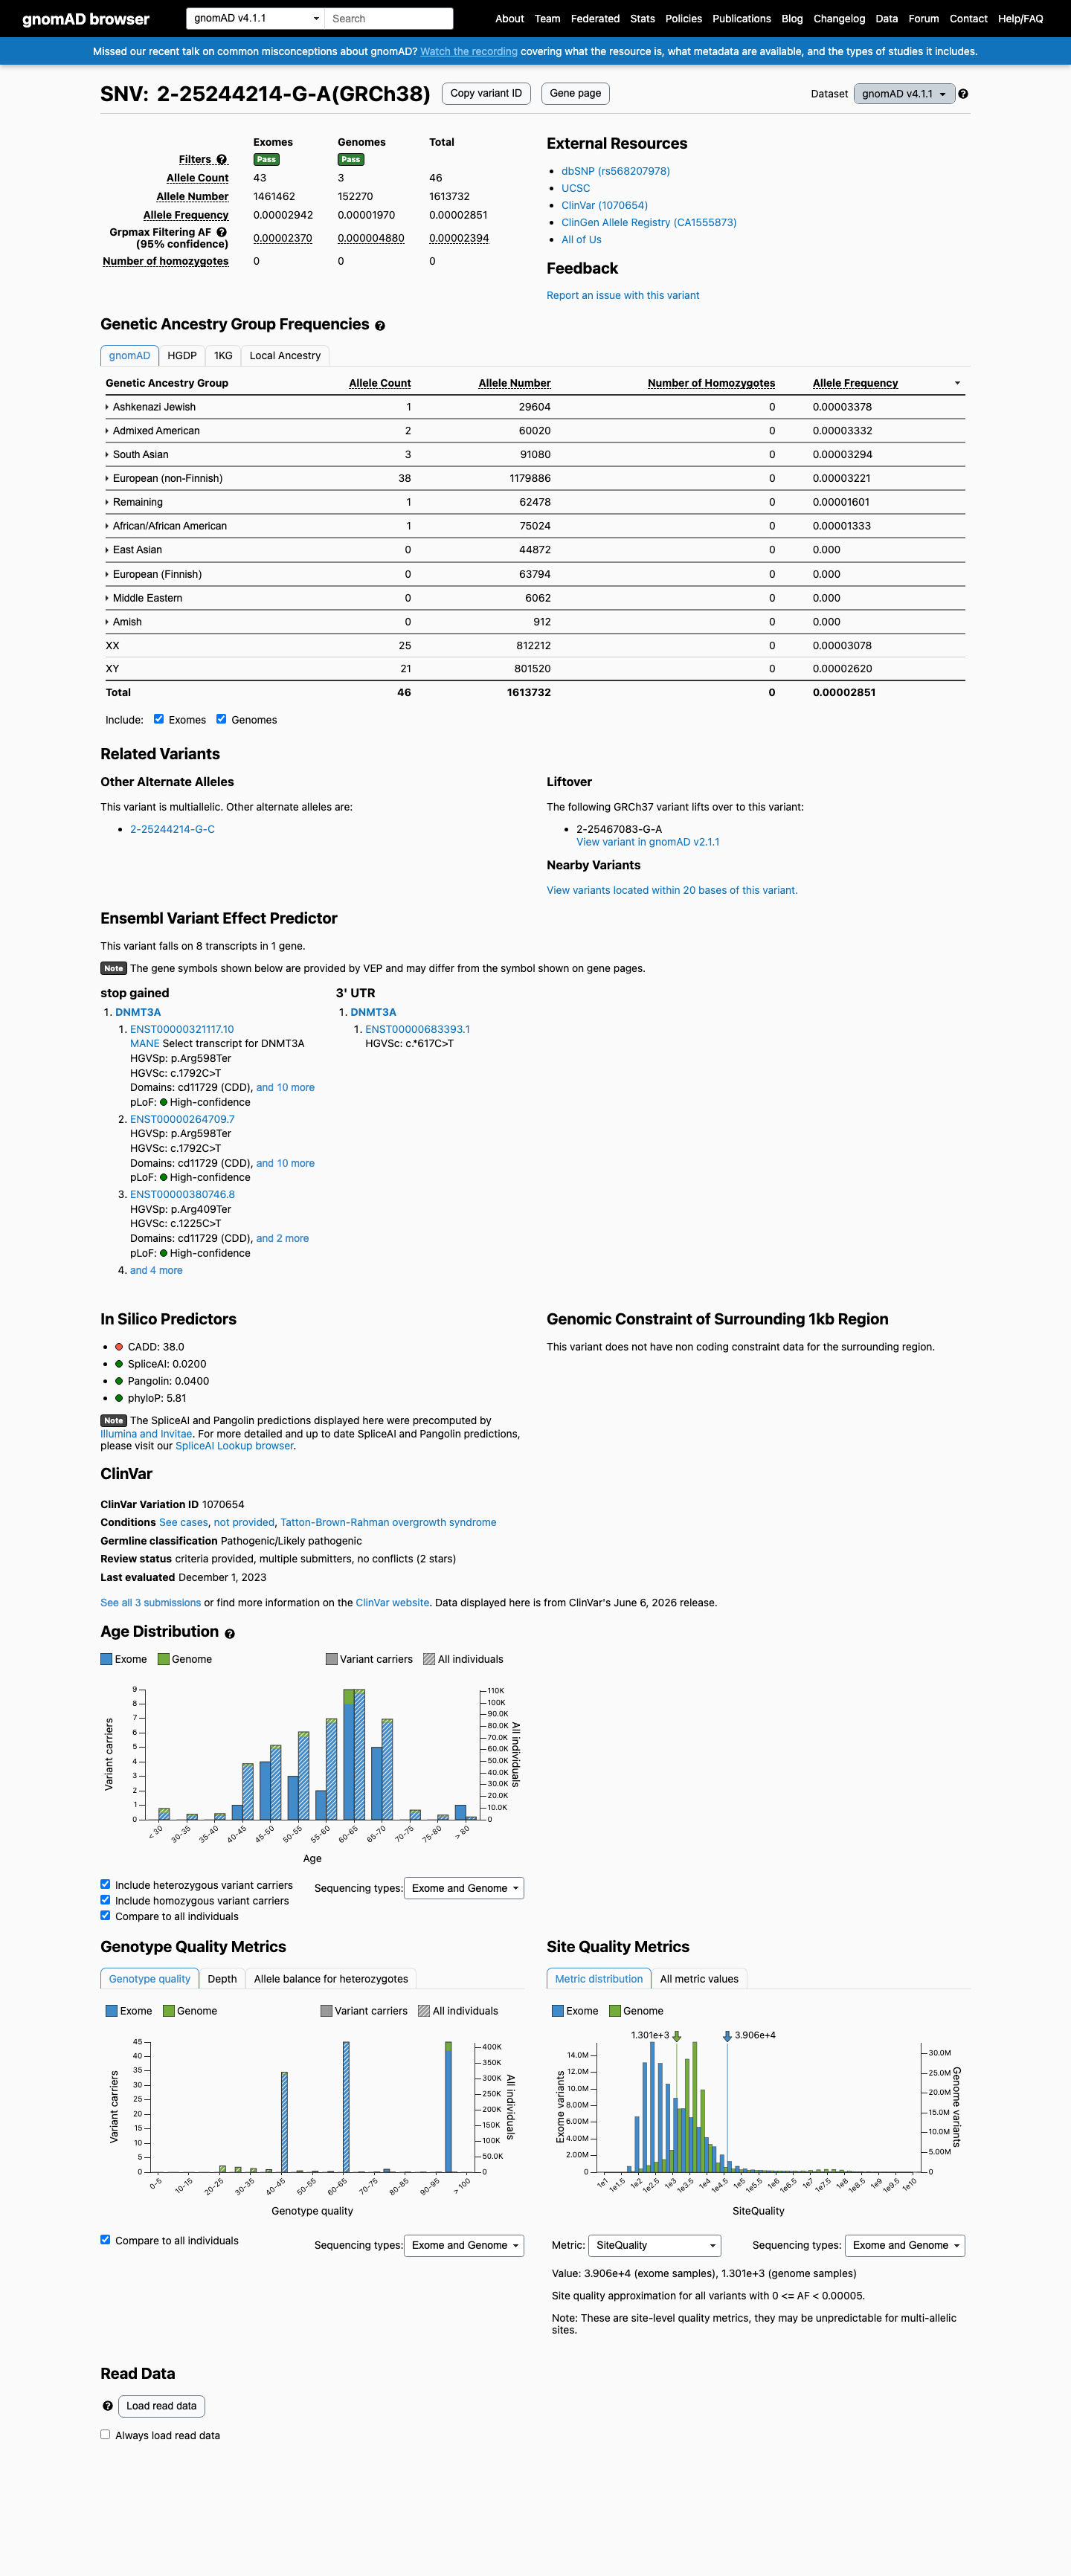
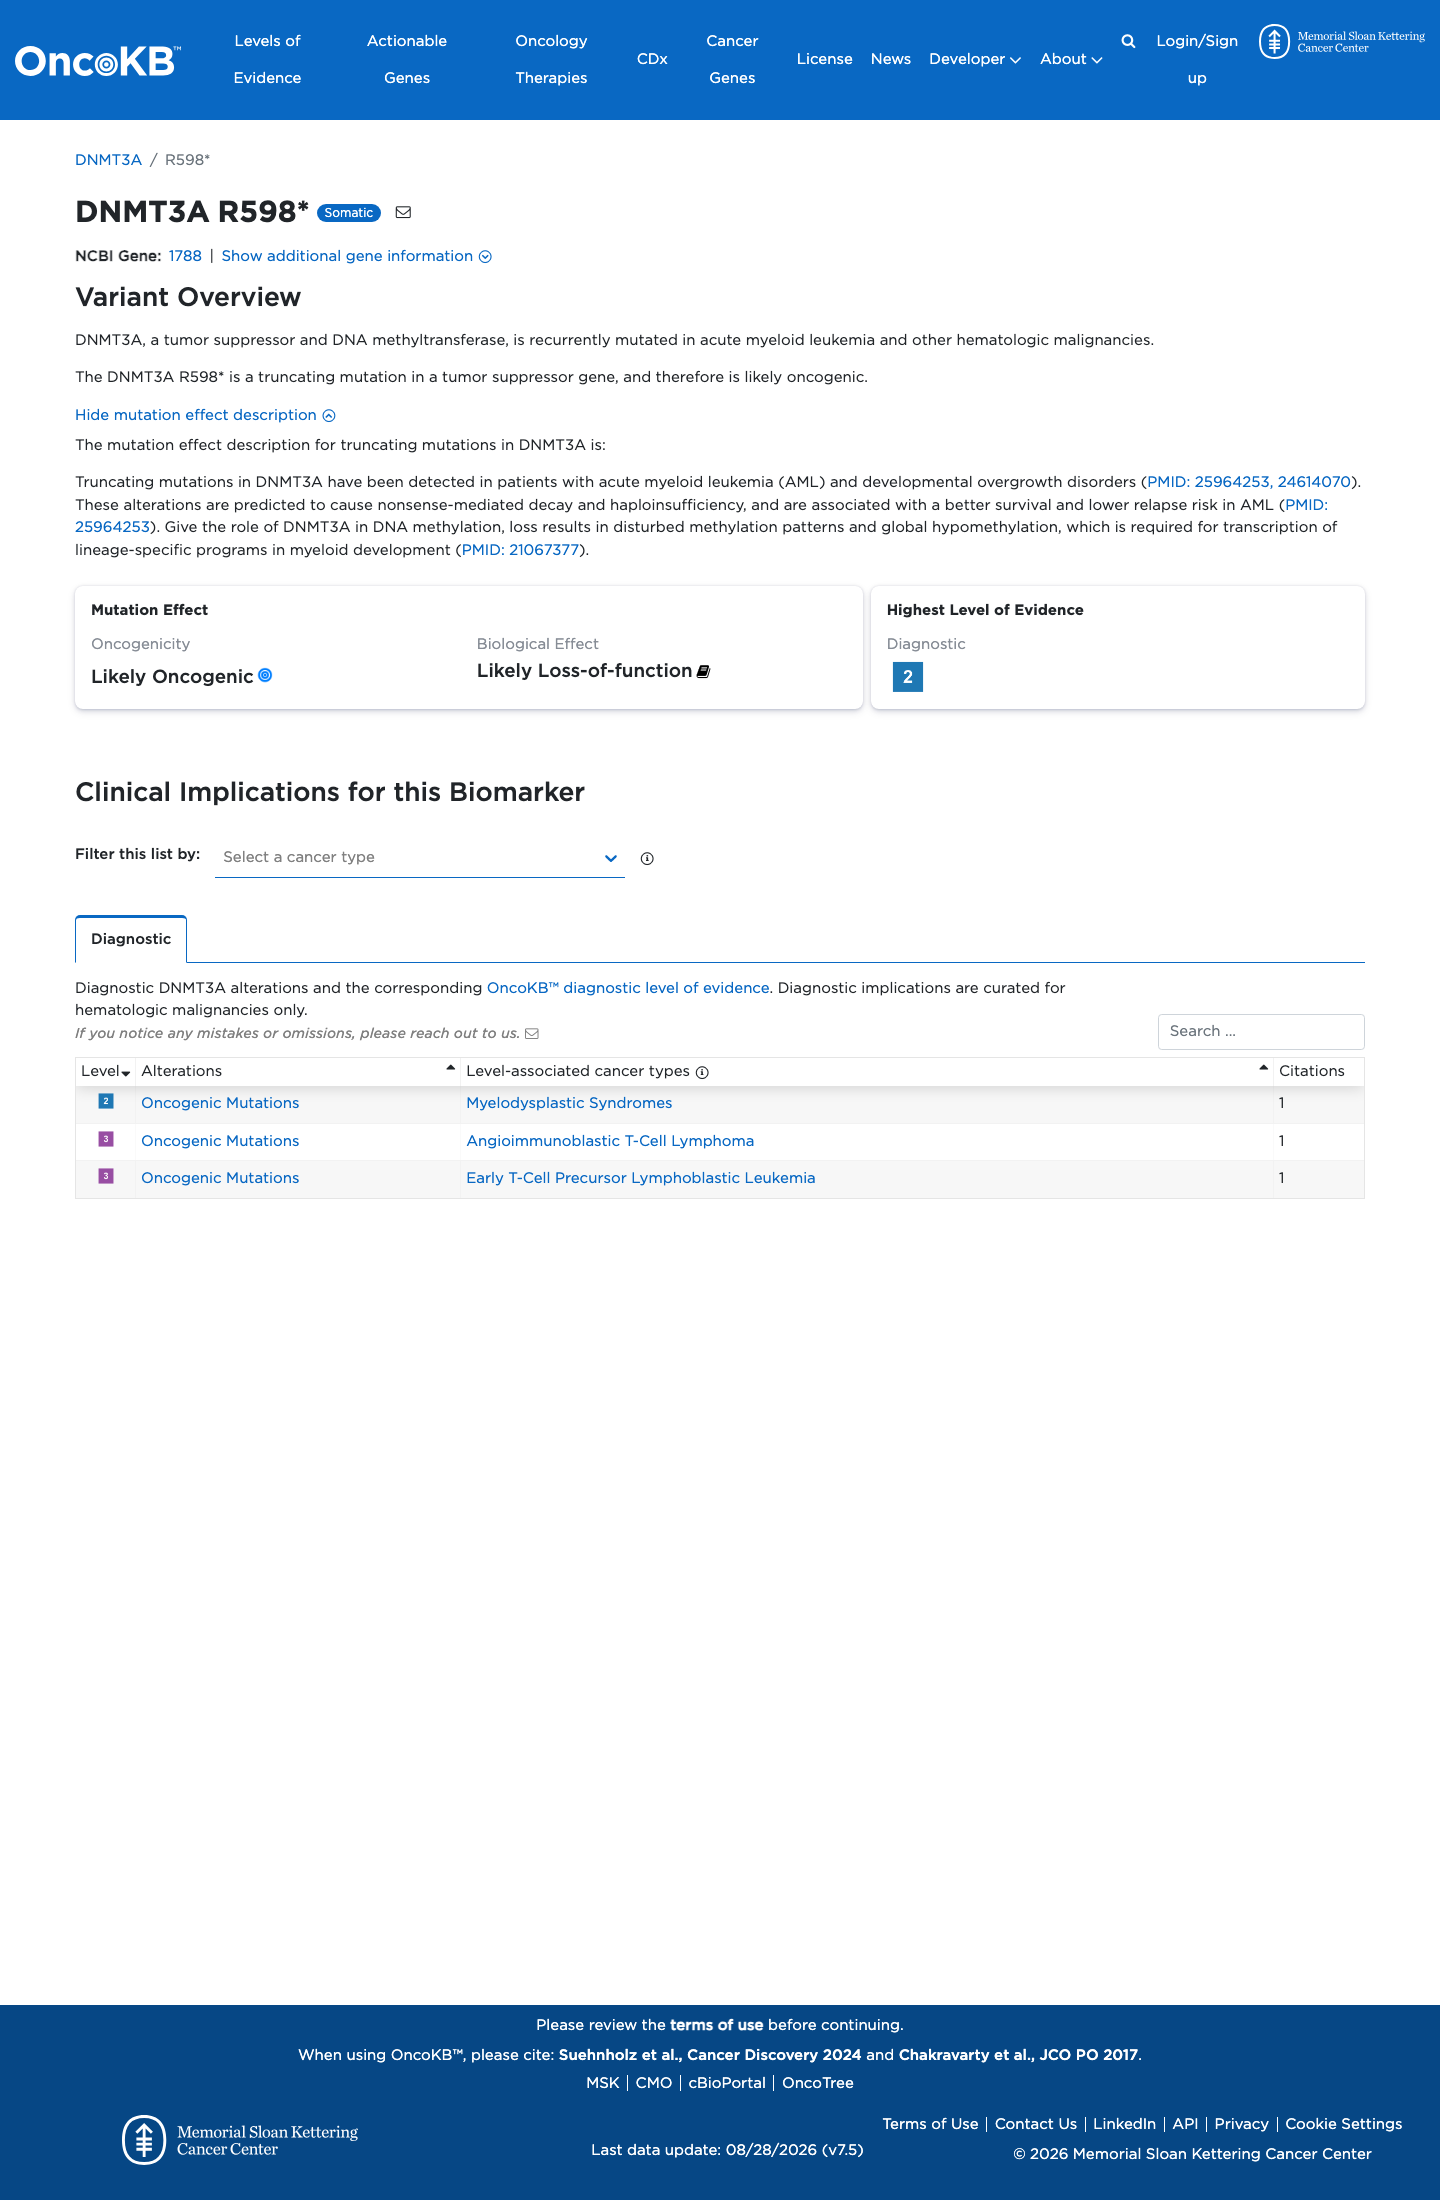
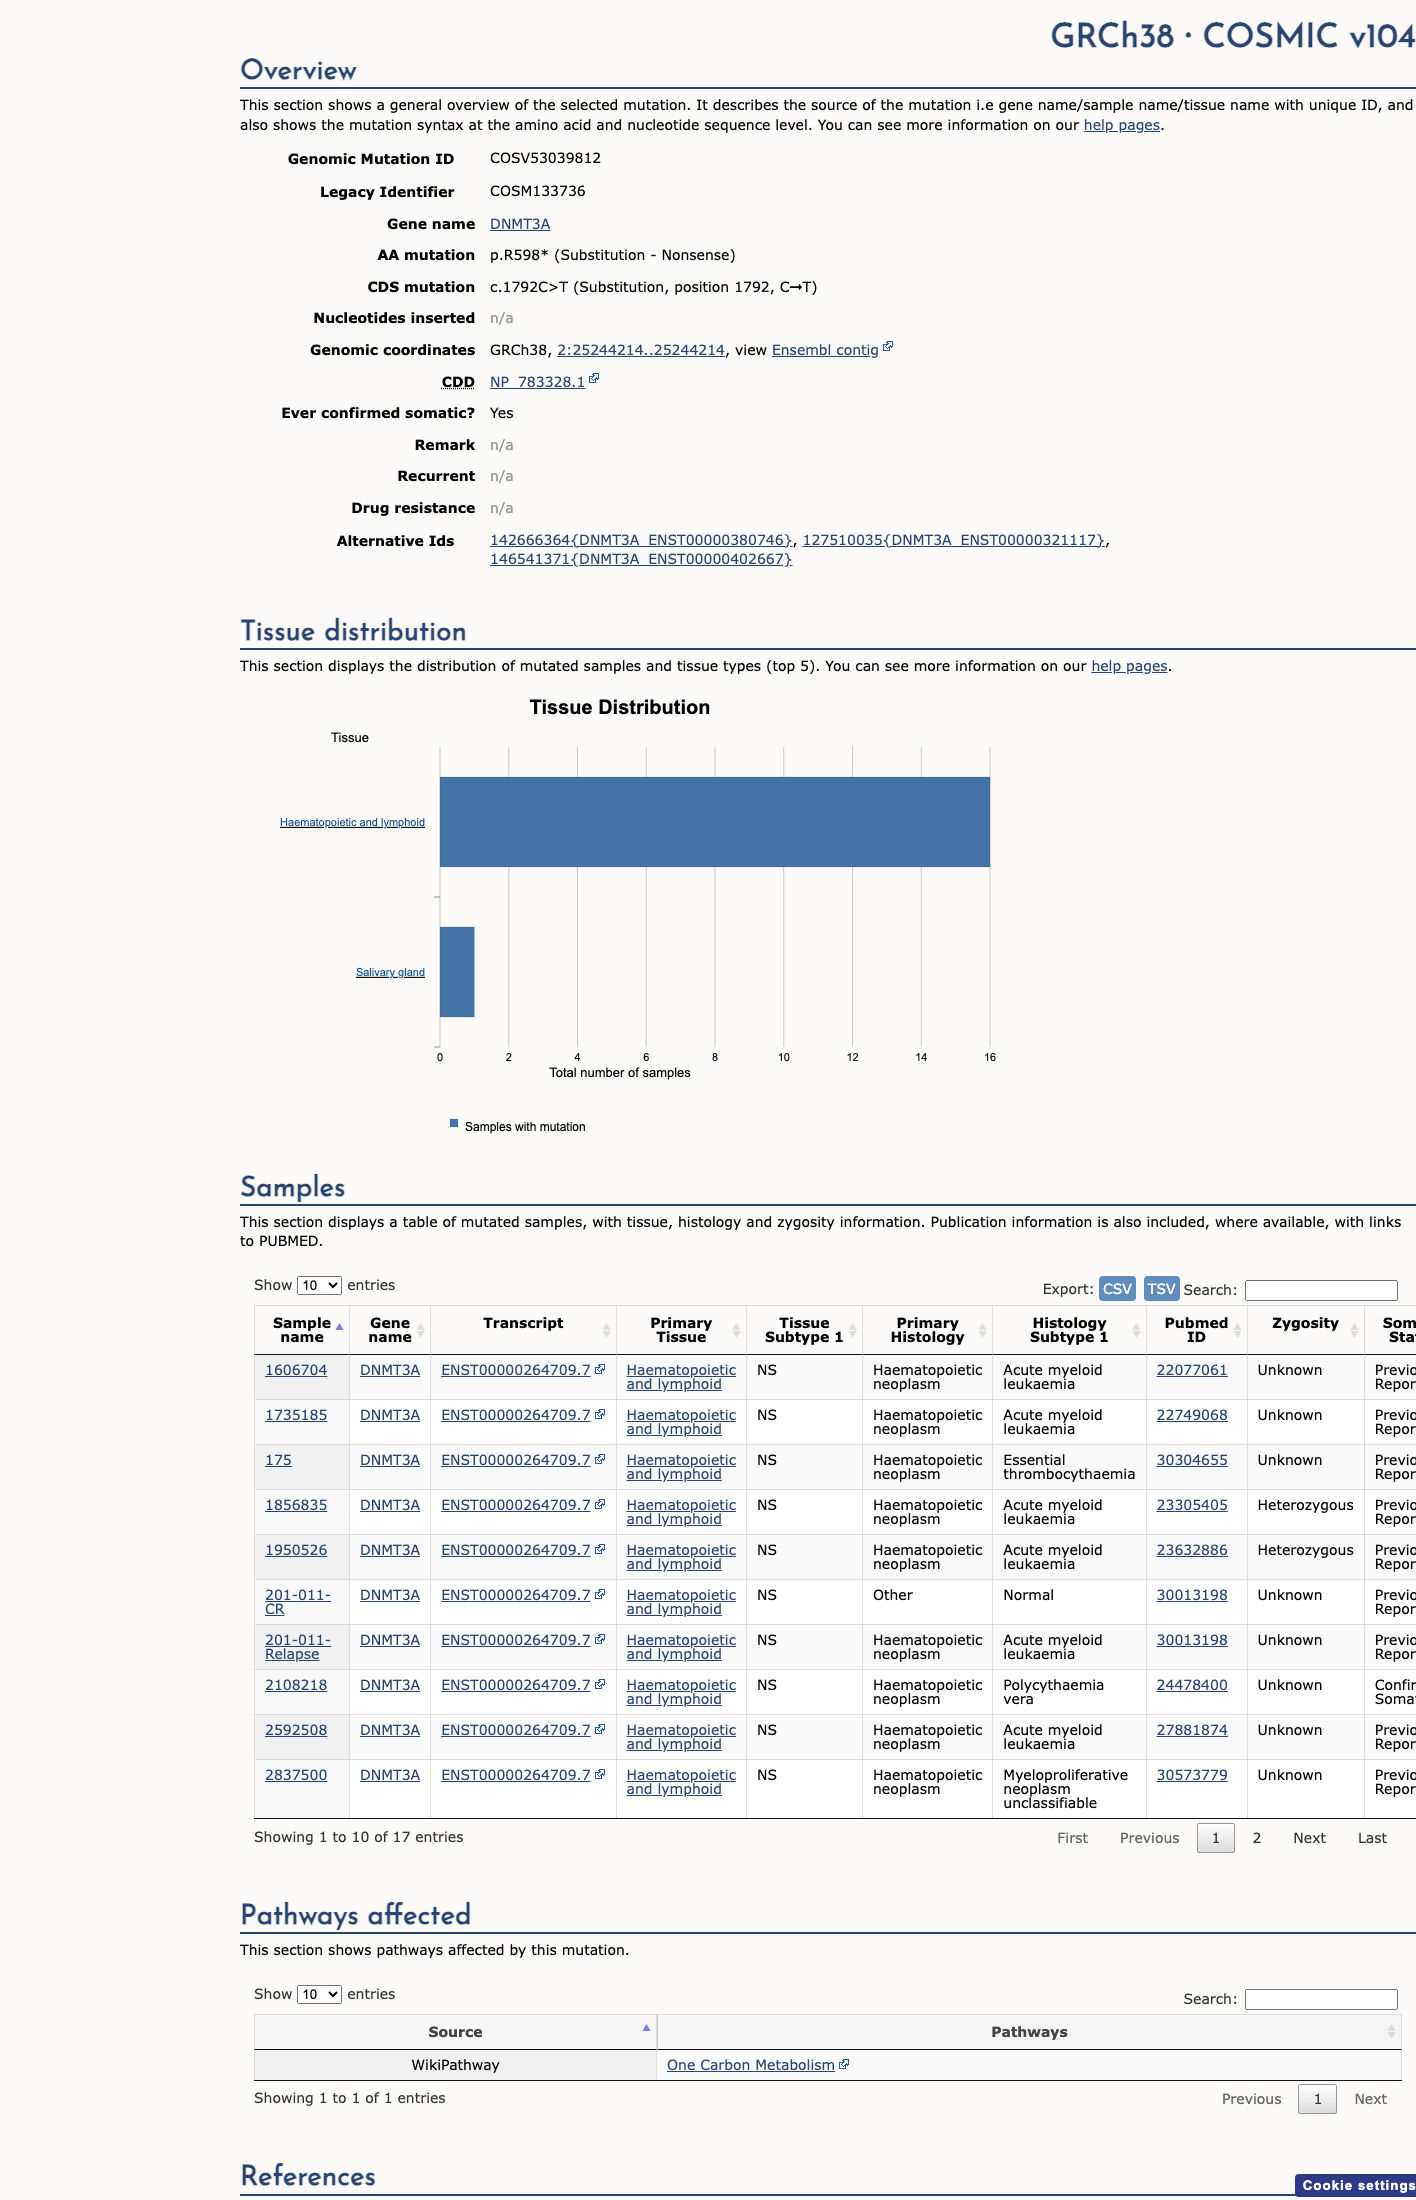
PVS1 (very strong): the premature stop at p.Arg598Ter is predicted to trigger nonsense-mediated decay. PM2 (supporting): the variant is rare in gnomAD, with AF 2.85054e-05 and zero homozygotes.
DNMT3A encodes an enzyme that adds new DNA methylation marks, helping control which genes are active during development and maintain normal genomic regulation. Germline changes in DNMT3A can cause Tatton-Brown-Rahman overgrowth syndrome, which includes intellectual disability. DNMT3A is also frequently altered in blood cancers, especially acute myeloid leukemia, and acts as a tumor suppressor in cancer.
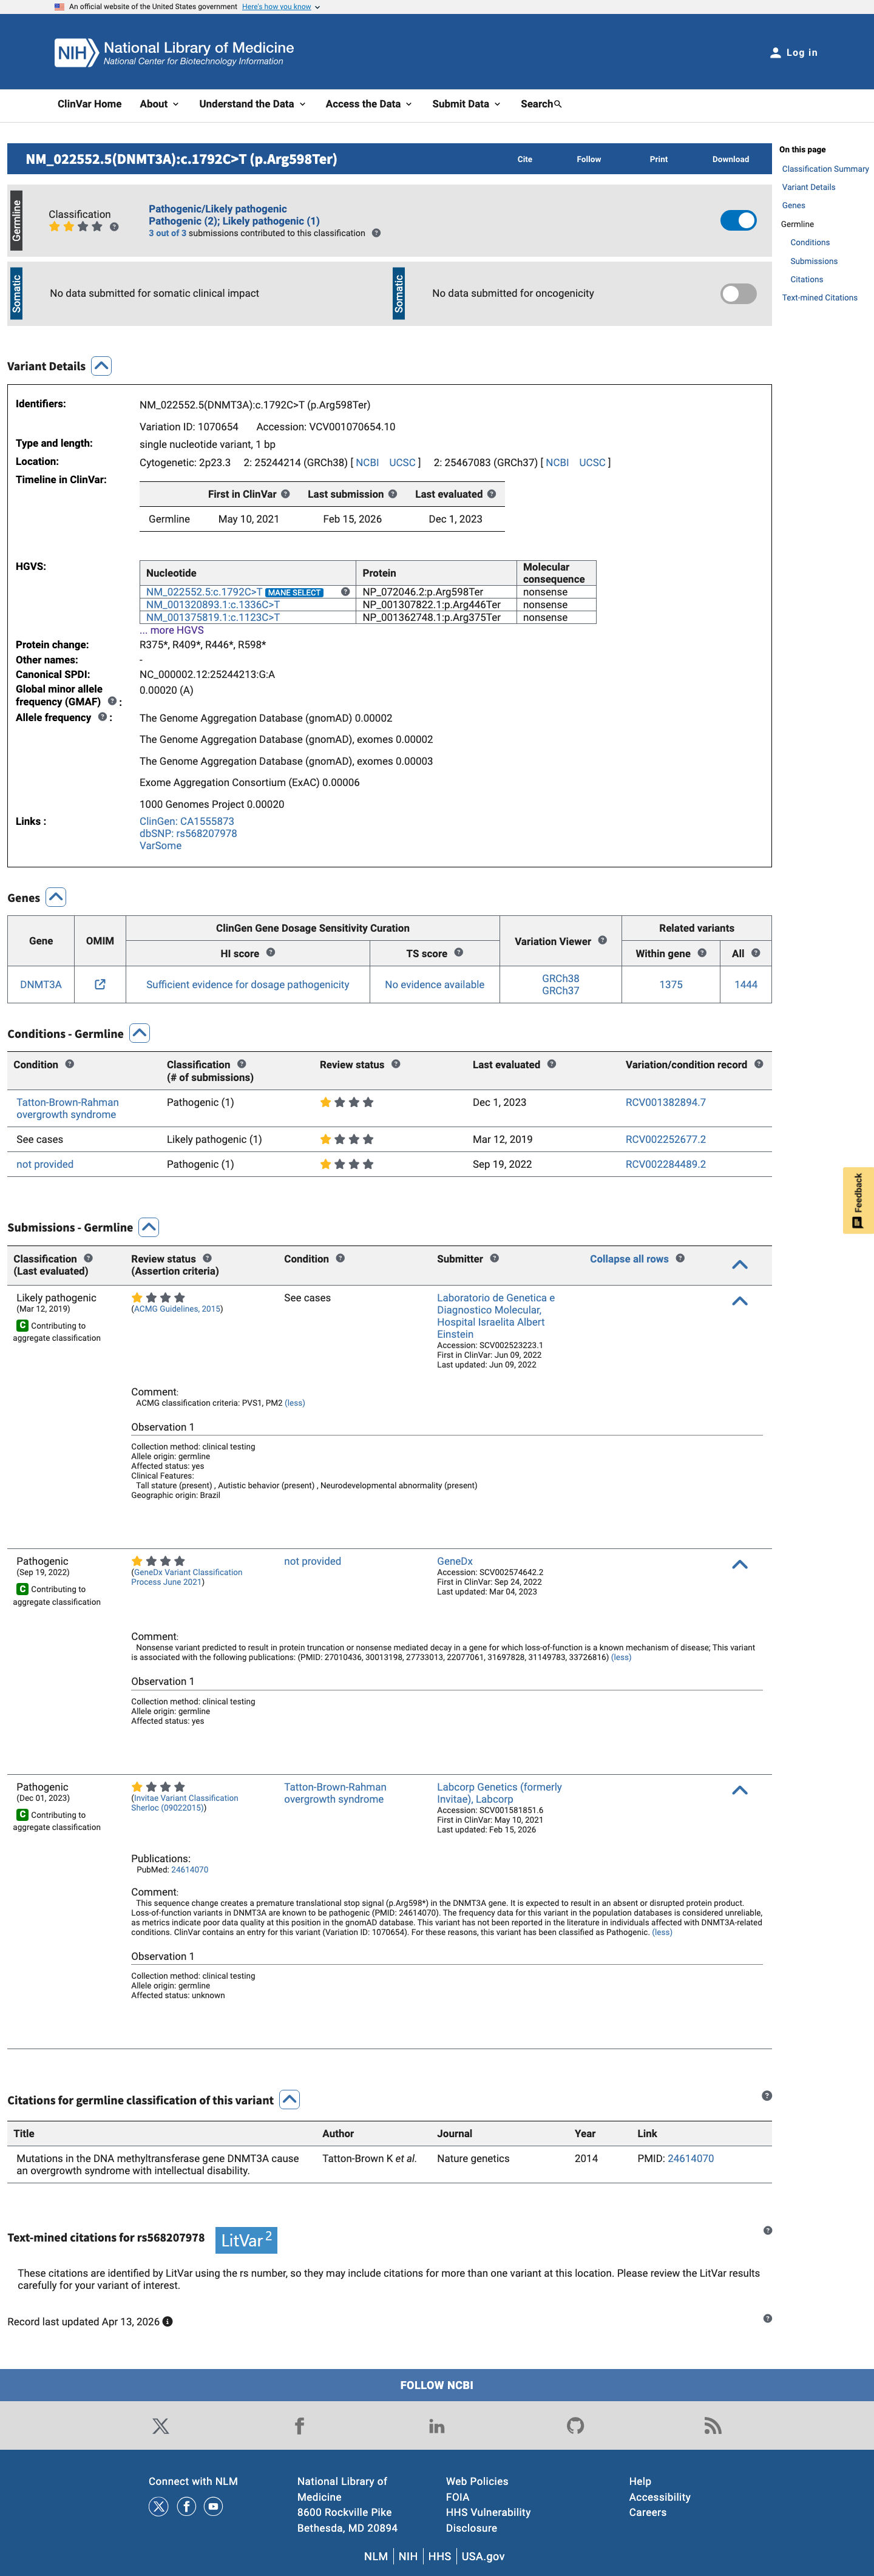
This truncating DNMT3A variant is relevant to the gene's established loss-of-function role in Tatton-Brown-Rahman overgrowth syndrome and its broader role in genomic regulation.
PVS1 (very strong): the premature stop at p.Arg598Ter is predicted to trigger nonsense-mediated decay. PM2 (supporting): the variant is rare in gnomAD, with AF 2.85054e-05 and zero homozygotes.
Ashkenazi Jewish 1 / 29,604 |
0.0034% |
Admixed American 2 / 60,020 |
0.0033% |
South Asian 3 / 91,080 |
0.0033% |
European (non-Finnish) 38 / 1,179,886 |
0.0032% |
Remaining individuals 1 / 62,478 |
0.0016% |
African/African American 1 / 75,024 |
0.0013% |
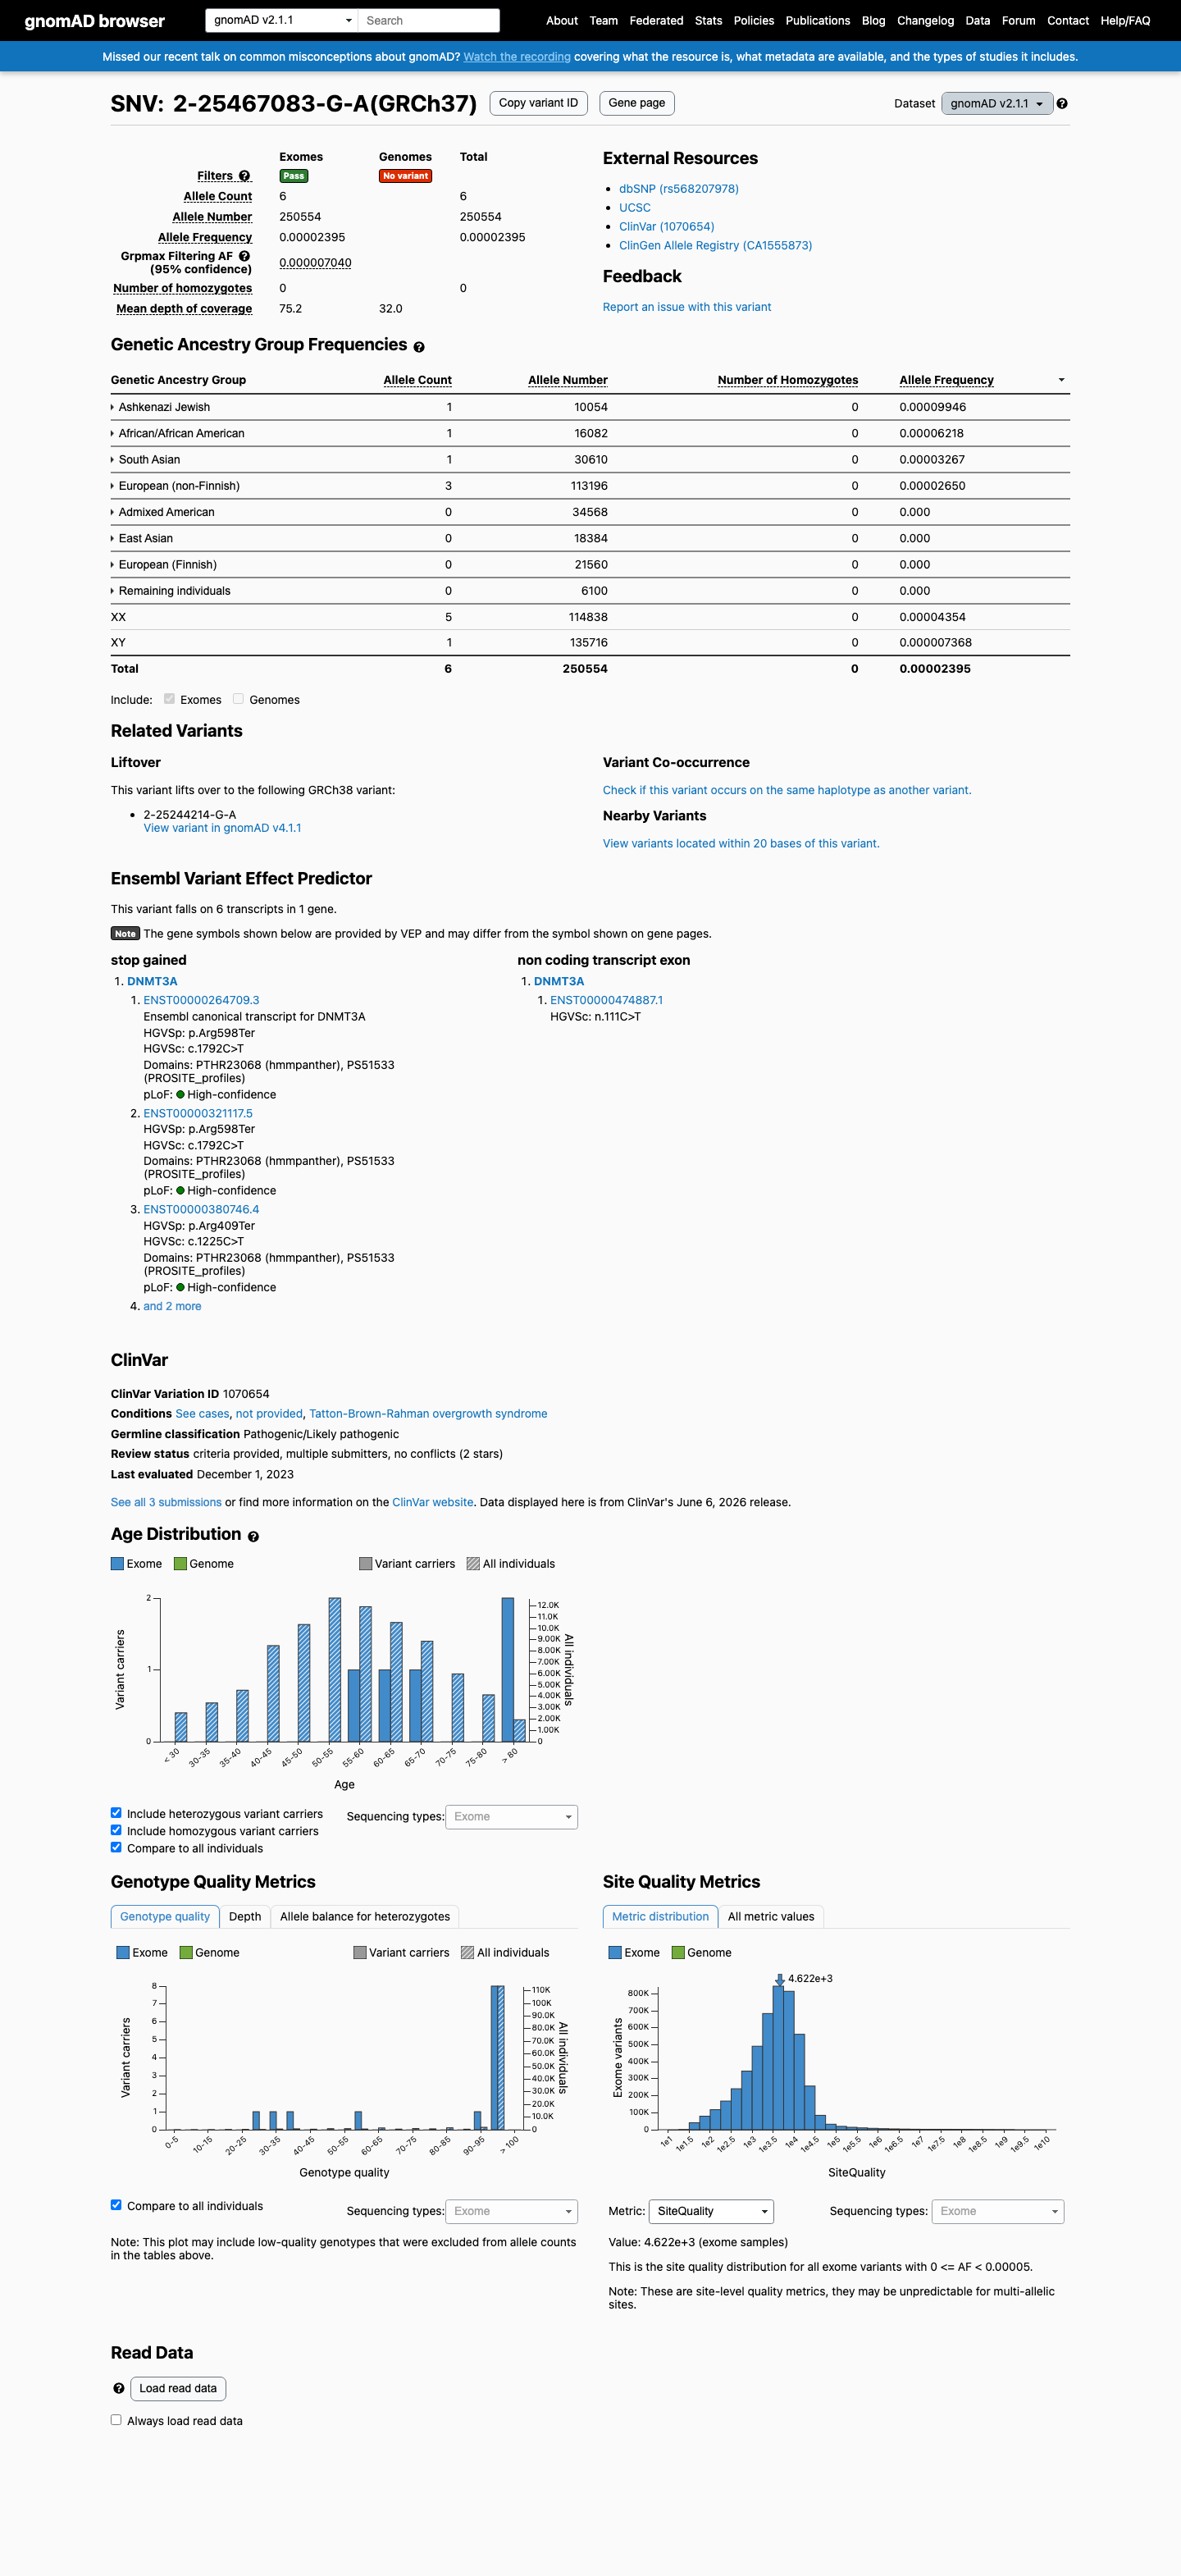
Ashkenazi Jewish 1 / 10,054 |
0.0099% |
African/African American 1 / 16,082 |
0.0062% |
South Asian 1 / 30,610 |
0.0033% |
European (non-Finnish) 3 / 113,196 |
0.0027% |