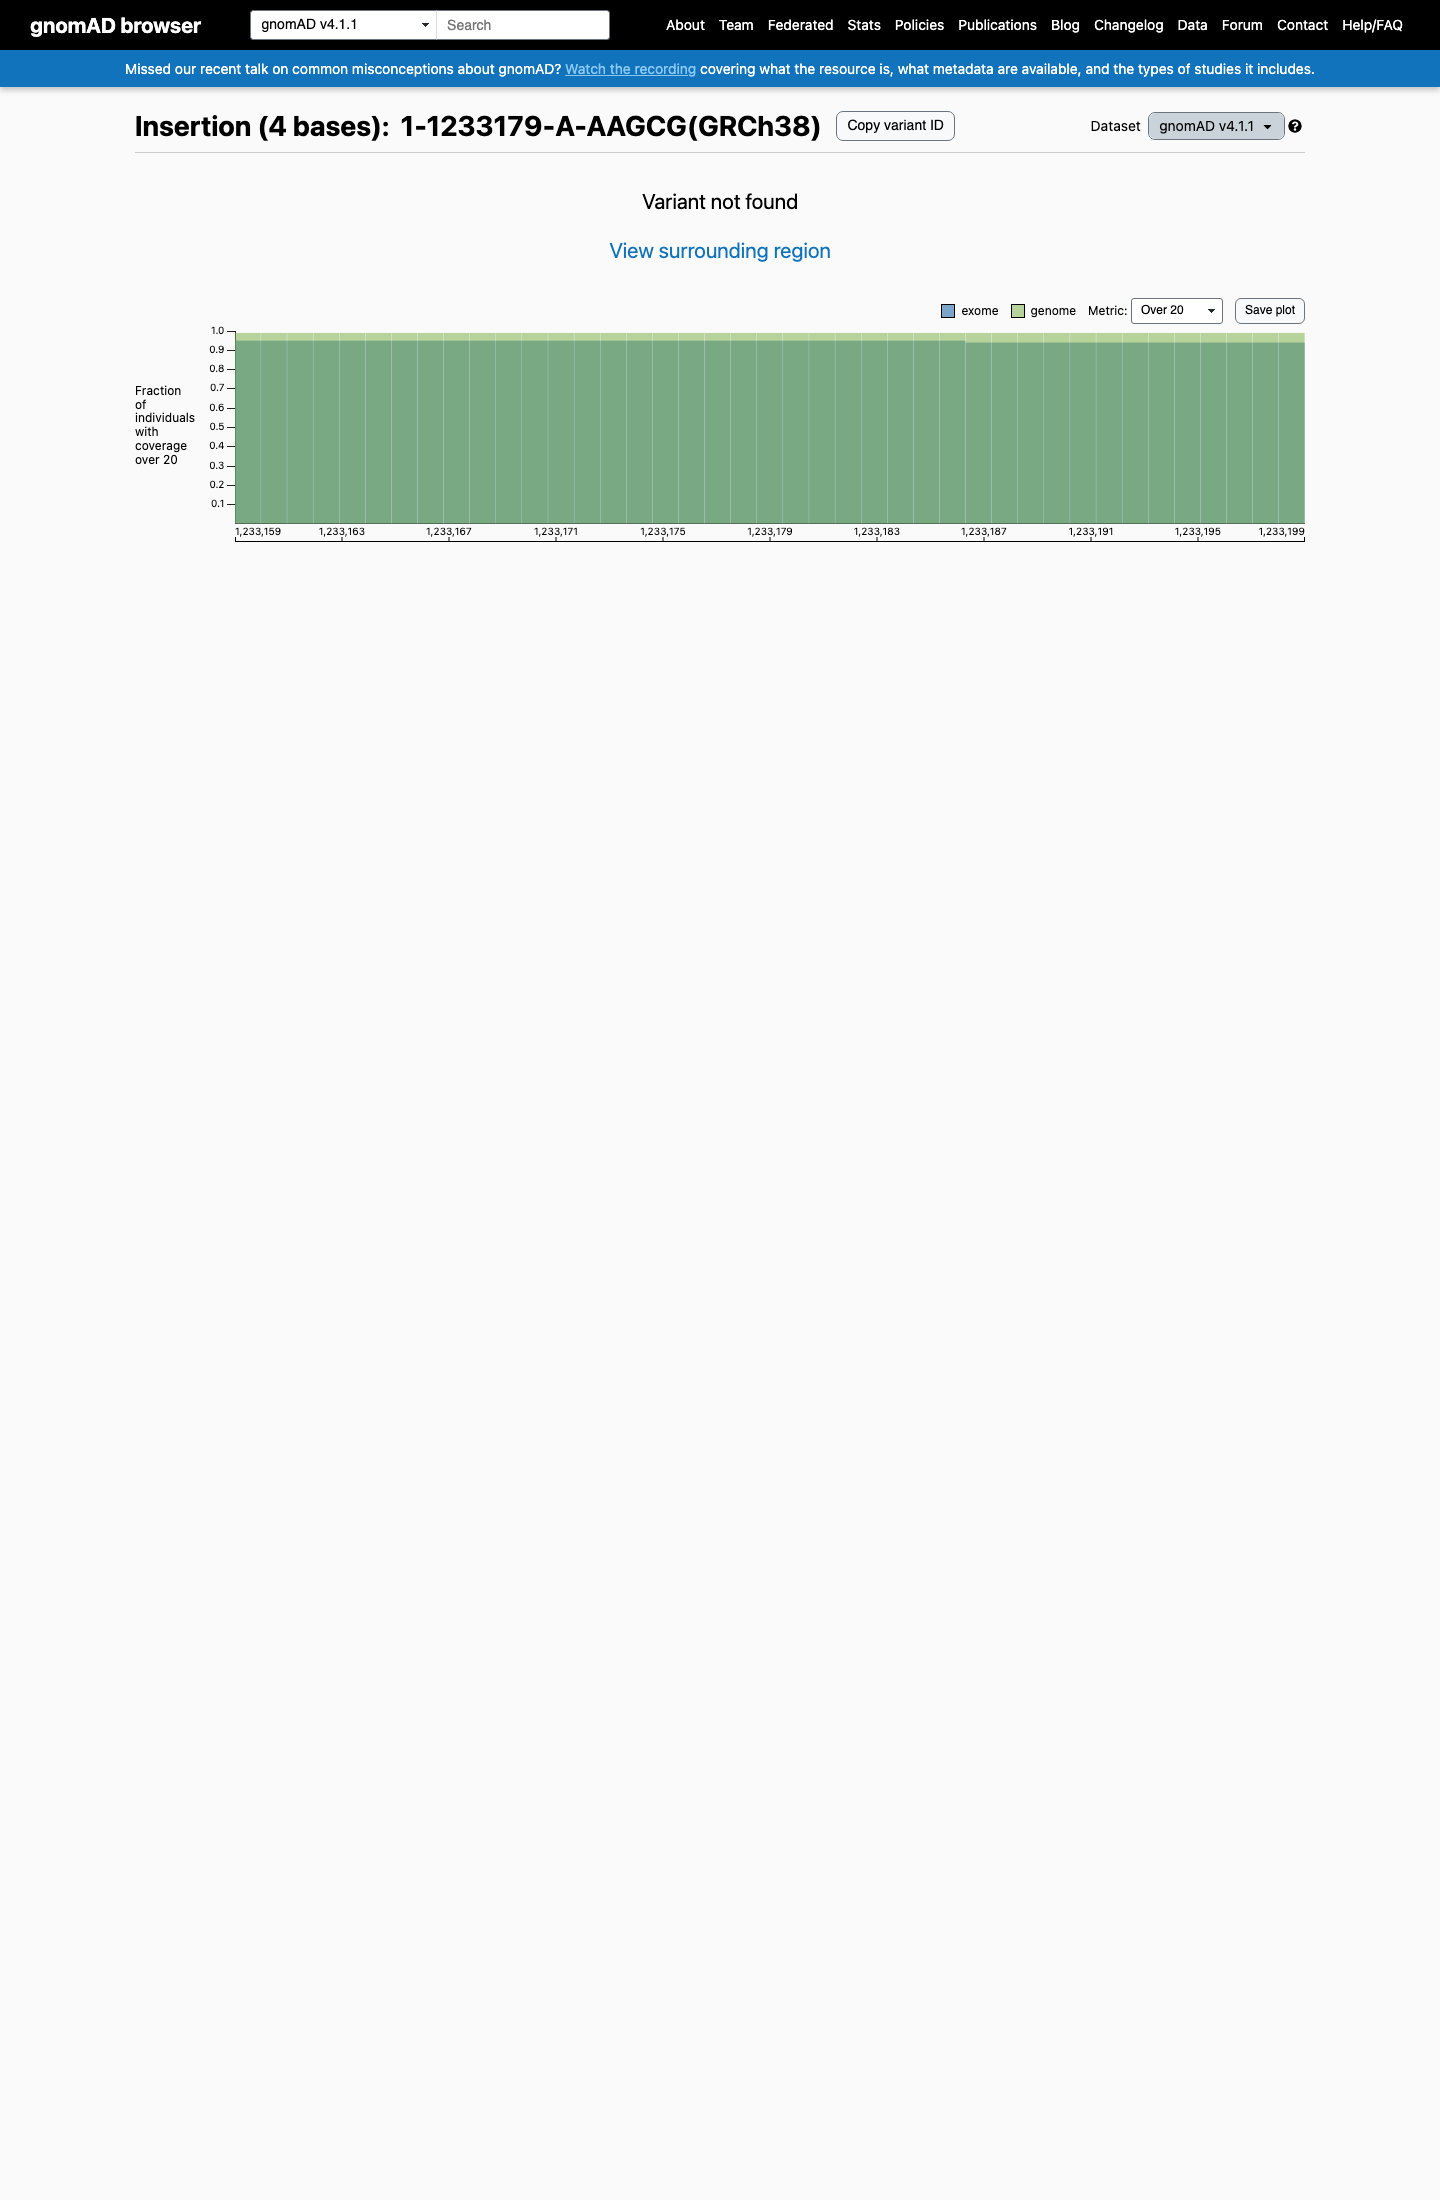
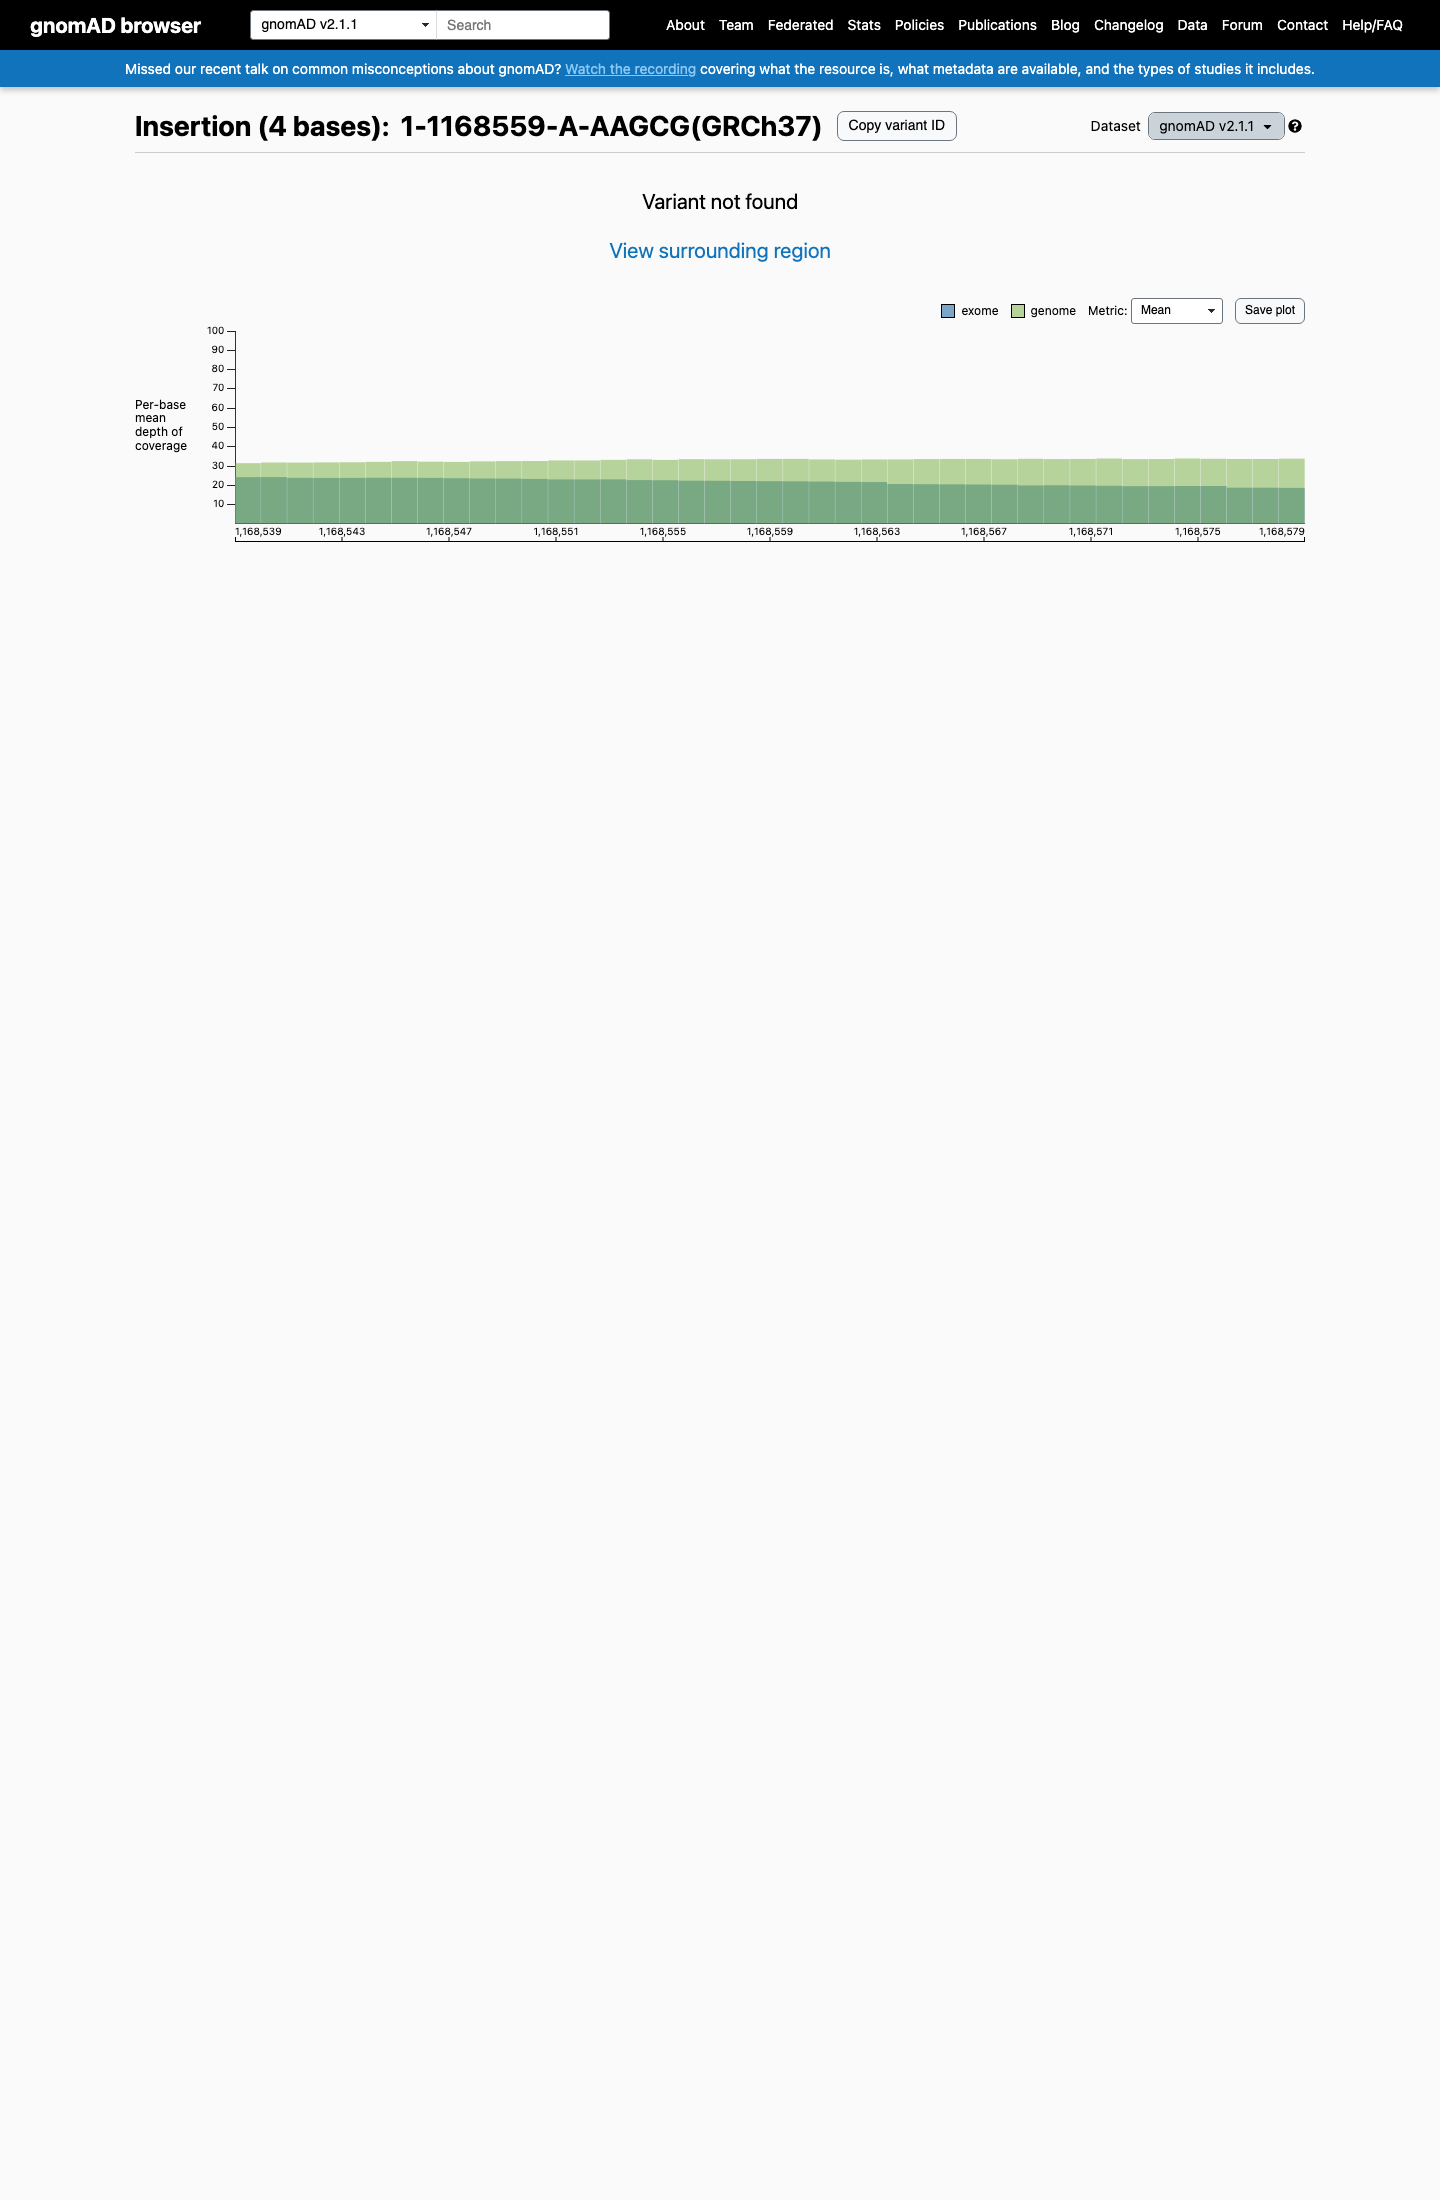
NM_080605.4:c.902_905dup is a 4bp duplication producing a frameshift predicted to yield NP_542172.2:p.(Glu303AlafsTer141) in B3GALT6, a gene where loss of function is an established disease mechanism for autosomal recessive Ehlers-Danlos-syndrome-like connective tissue disorder (PVS1_Moderate; PMID:23664118, PMC6185798). The NM_080605.4 transcript is MANE Select and represents the single exon of B3GALT6; nonsense-mediated decay is not expected, warranting a downgrade from PVS1_VeryStrong to PVS1_Moderate per ClinGen SVI PVS1 recommendations.1 This variant is absent from gnomAD v2.1, gnomAD v4.1, and gnomAD-Canada v1.0 population databases (allele frequency = 0.0 in all datasets), meeting PM2 for extremely low population frequency.2 Combining the above, two moderate-level pathogenic criteria (PVS1_Moderate + PM2_Moderate) are insufficient to reach a Likely Pathogenic classification under generic ACMG/AMP 2015 combination rules (PMID:25741868), which require at minimum one Very Strong plus one Moderate, one Strong plus one Moderate, or three Moderate criteria. No benign criteria are met. The variant is classified as a Variant of Uncertain Significance (VUS).3
Final classification
VUS
PVS1PM2
B3GALT6
c.902_905dup
p.Glu303AlafsTer141
This variant
NM_080605.4:c.902_905dup is a 4bp duplication producing a frameshift predicted to yield NP_542172.2:p.(Glu303AlafsTer141) in B3GALT6, a gene where loss of function is an established disease mechanism for autosomal recessive Ehlers-Danlos-syndrome-like connective tissue disorder (PVS1_Moderate; PMID:23664118, PMC6185798). The NM_080605.4 transcript is MANE Select and represents the single exon of B3GALT6; nonsense-mediated decay is not expected, warranting a downgrade from PVS1_VeryStrong to PVS1_Moderate per ClinGen SVI PVS1 recommendations.
gene-specific framework lacked a usable explicit final combination framework, so generic ACMG/AMP 2015 final-combination rules were applied as fallback; applied criteria: PVS1 moderate, PM2 moderate; combination = 2 moderate, which maps to VUS.
Classification rationale
PVS1PM2
VUS
B3GALT6 c.902_905dup
PVS1 + PM2
→
VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria
may be applied incorrectly, sources may be misread, and a confident-looking
classification can still be wrong. Double-check every criterion and
its underlying evidence before relying on any call.
Gene diagram
· NM_080605.4 · variants mapped to exon structure
B3GALT6
NM_080605.4
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 18 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength
Supporting
Moderate
Strong
Very strong
PVS1
moderate
Pathogenic
NM_080605.4:c.902_905dup is a 4bp duplication producing a frameshift that predicts NP_542172.2:p.(Glu303AlafsTer141). B3GALT6 loss of function is an established disease mechanism for autosomal recessive Ehlers-Danlos-syndrome-like connective tissue disorder (PMID:23664118). The NM_080605.4 transcript is MANE Select and constitutes the single exon of B3GALT6; nonsense-mediated decay is not expected. Under ClinGen SVI PVS1 recommendations (PMC6185798), frameshift variants in single-exon genes where NMD cannot occur warrant a downgrade from Very Strong to Moderate.
Frameshift variant predicted to yield NP_542172.2:p.(Glu303AlafsTer141)B3GALT6 LoF established as disease mechanism for autosomal recessive EDS-like connective tissue disorderNM_080605.4 is MANE Select
PM2
moderate
Pathogenic
NM_080605.4:c.902_905dup is absent from gnomAD v2.1, gnomAD v4.1, and gnomAD-Canada v1.0 population databases (allele frequency = 0.0 in all datasets), meeting PM2 for extremely low/absent population frequency.
Absent from gnomAD v2.1 (0 alleles)Absent from gnomAD v4.1 (0 alleles)Absent from gnomAD-Canada v1.0 (0 alleles)
Assessed · not applied
· 4 not met · 14 not assessed
Pathogenic
PS2
No de novo data with confirmed paternity and maternity available for this variant.
PS3
No well-established in vitro or in vivo functional studies specific to NM_080605.4:c.902_905dup are available.
PS4
No case-control data demonstrating statistically increased prevalence of this variant in affected individuals versus controls is available.
PM1
No data demonstrating that this variant lies within a well-established mutational hotspot or critical functional domain with supporting statistical evidence is available.
PM6
No assumed de novo observation (without parental confirmation) is available for this variant.
PP1
No cosegregation data in multiple affected family members is available for this variant.
PP3
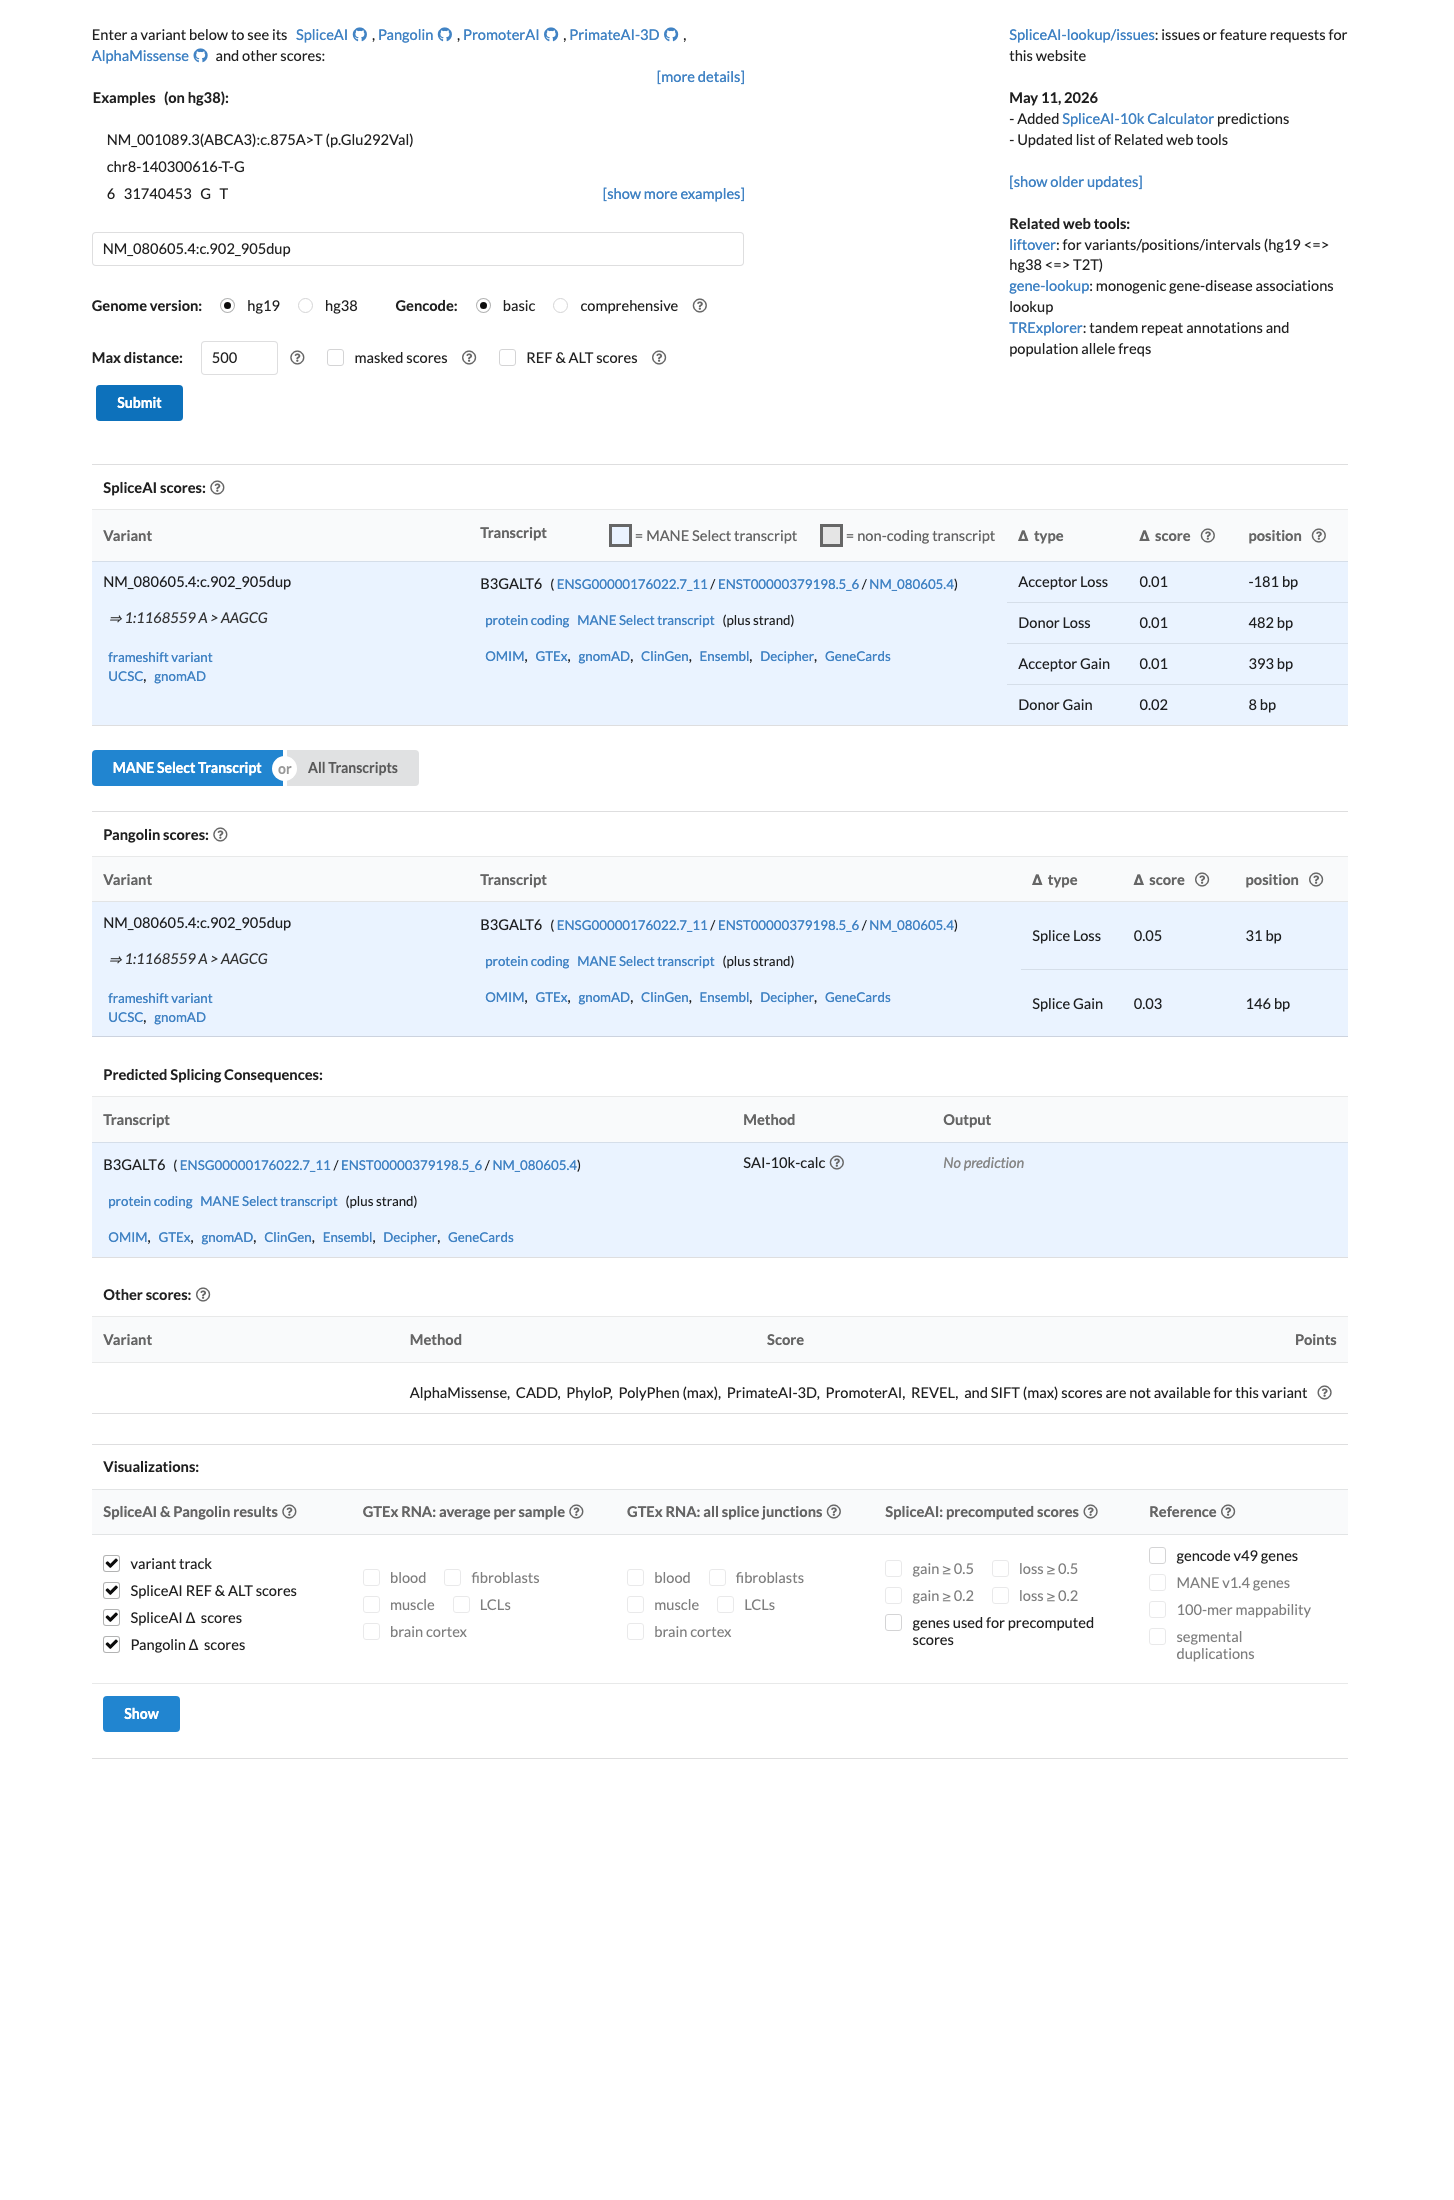
SpliceAI predicts no significant splice impact (max delta score = 0.02).
PP4
No patient phenotype or family history data specific to this variant is available.
PP5
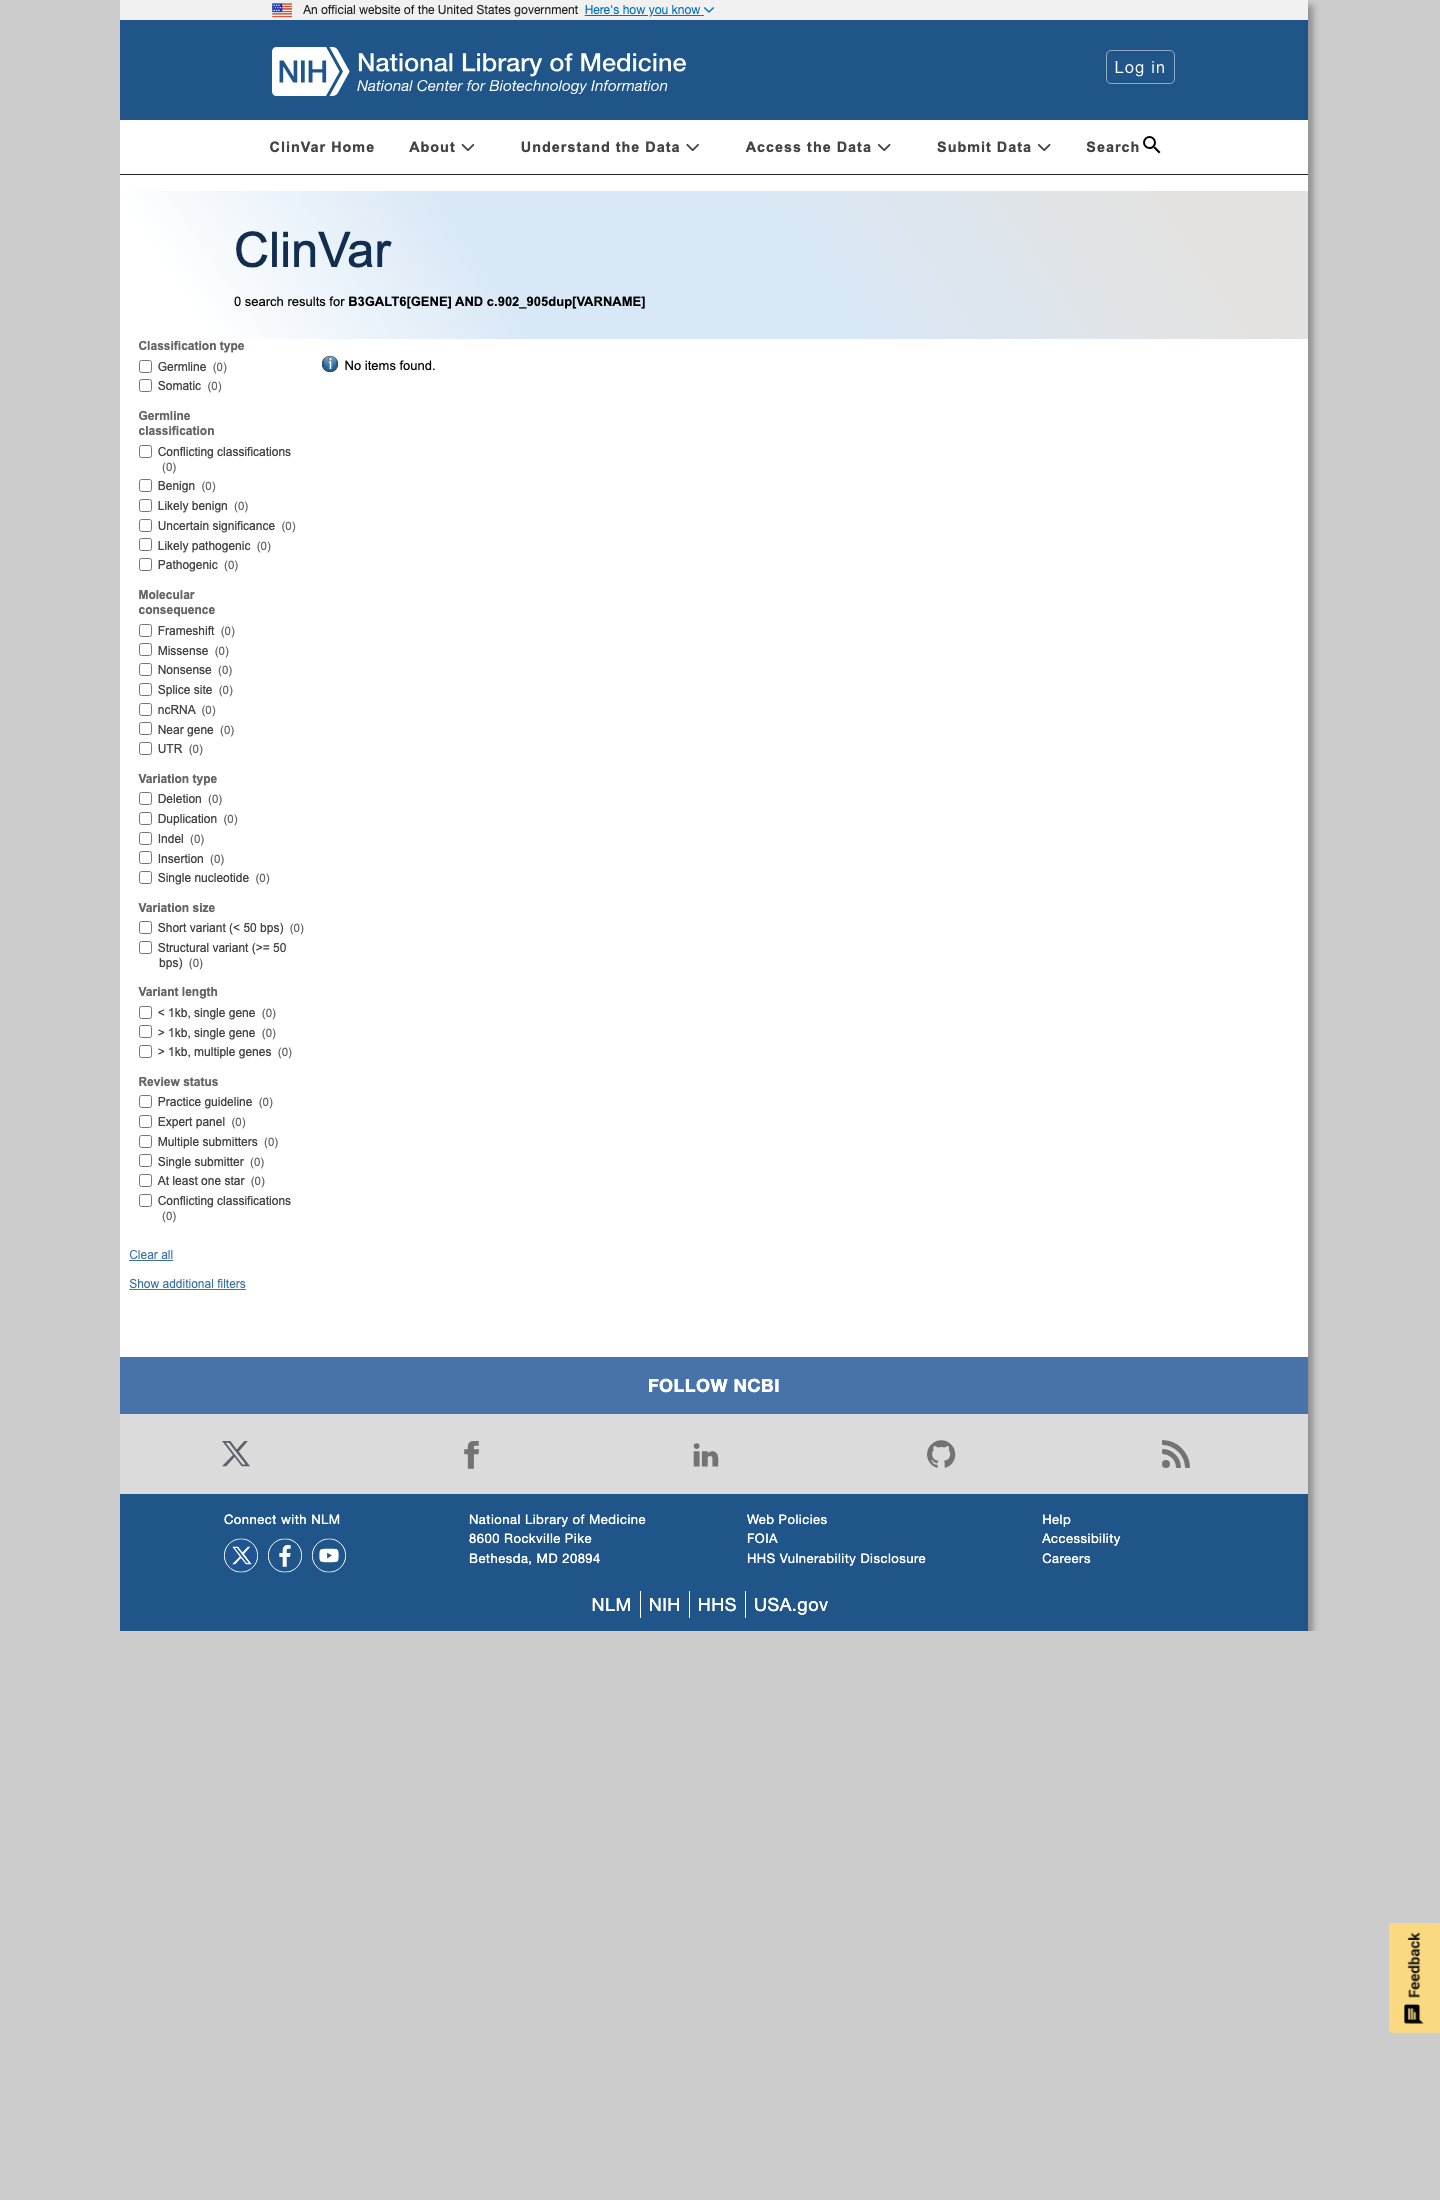
This variant is absent from ClinVar; no reputable source has reported it as pathogenic.
Benign
BA1
This variant is absent from gnomAD v2.1, v4.1, and gnomAD-Canada (allele frequency = 0.0), far below the BA1 threshold of >1%.
BS1
This variant is absent from all population databases (AF = 0.0), failing to meet the BS1 threshold of >0.3% allele frequency.
BS2
No data on observation of this variant in healthy adult individuals for a fully penetrant disorder is available.
BS3
No well-established in vitro or in vivo functional studies demonstrating no damaging effect for this specific variant are available.
BS4
No segregation data demonstrating lack of cosegregation with disease in affected family members is available.
BP2
No data on observation of this variant in trans with a pathogenic variant (for a dominant disorder) or in cis with a pathogenic variant (any inheritance pattern) is available.
BP4
Multiple lines of computational evidence do not support a benign effect.
BP5
No data on an alternate molecular basis for disease in a case carrying this variant is available.
BP6
This variant is absent from ClinVar; no reputable source has reported it as benign.
N/A · 8
PS1 · PM3 · PM4 · PM5 · PP2 · BP1 · BP3 · BP7
Research & evidence
Population frequency
gnomAD v4.1
gnomAD v2.1
v4.1
Absent from gnomAD v4.1.
v2.1
Absent from gnomAD v2.1.
🇨🇦 CA
This variant is absent from gnomAD-Canada.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
Absent
· 0 / ?
0 hom
0 hom
Not observed in any ancestry group.
gnomAD v2.1
Absent
· 0 / ?
0 hom
0 hom
Not observed in any ancestry group.
gnomAD Canada 🇨🇦
Absent
· 0 / ?
0 hom
0 hom
Not observed in any ancestry group.
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.02).
Functional
No data
No calibrated functional assay or RNA evidence was identified for this variant.

COSMIC
Cancer hotspots
Somatic evidence
Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Literature · how each cited paper was used
1papers cited
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why.
PMID 23664118
Found
Structured finding pending for this record — see source link.
Applied to
→PVS1
moderate
Sources & reference links