Back
NM_000179.3:c.187T>C
p.Ser63Pro · MSH6
0%
complete
Final classification
Likely Benign
BS1BP4
MSH6
c.187T>C
p.Ser63Pro
missense · exon 1

MSH6 encodes a protein in the DNA mismatch repair system, which fixes errors made during DNA replication. Partnering with MSH2, it forms a complex that recognizes and helps correct mismatched DNA bases, keeping the genetic code stable. Inherited mutations in MSH6 cause Lynch syndrome (hereditary nonpolyposis colorectal cancer), raising the risk of colorectal, endometrial, ovarian, and other cancers, while mutations in both copies lead to constitutional mismatch repair deficiency. Because faulty mismatch repair drives tumor development and produces microsatellite instability, MSH6 acts as a tumor suppressor, and cancers with such repair defects often respond well to immune checkpoint inhibitor therapy.

This variant

MSH6 encodes a DNA mismatch-repair protein whose inherited disruption causes Lynch syndrome and, when biallelic, constitutional mismatch repair deficiency.

Transcript
NM_000179.3
HGVS · transcript:coding
NM_000179.3:c.187T>C
GRCh38
chr2:47783420 T>C
GRCh37
chr2:48010559 T>C
Likely Benign: BS1 (strong) plus BP4 (supporting) satisfy Rule18 of the InSiGHT MSH6 Version 2.0 framework.
Classification rationale
BS1BP4 Likely Benign
MSH6 c.187T>C missense · exon 1

BS1 Strong: gnomAD v4.1 grpmax allele frequency of 0.00055895 meets the MSH6 frequency interval. BP4 Supporting: MSH6 HCI prior probability of pathogenicity is 0.0027, below the <0.11 threshold.

BS1 + BP4 → Likely Benign
LYFE Sciences is an AI system, and it can make mistakes. Criteria may be applied incorrectly, sources may be misread, and a confident-looking classification can still be wrong. Double-check every criterion and its underlying evidence before relying on any call.
Gene diagram · NM_000179.3 · variants mapped to exon structure
MSH6 NM_000179.3
Fetching transcript structure from UCSC…
Applied criteria · 2 applied · 14 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 2
Strength Supporting Moderate Strong Very strong
BS1 strong Benign
Met, strong: gnomAD v4.1 grpmax filtering allele frequency 0.00055895 falls between the MSH6 BS1 thresholds 0.00022 and 0.0022.
The MSH6 InSiGHT VCEP BS1 Strong rule requires gnomAD v4 grpmax filtering allele frequency ≥0.00022 and <0.0022 (0.022–0.22%) and exclusion as a founder pathogenic variant.gnomAD v4.1 reports grpmax filtering allele frequency 0.00055895 (0.055895%), which lies within the VCEP BS1 interval; no founder-pathogenic designation is documented.
BP4 supporting Benign
Met, supporting: MSH6 HCI prior probability 0.0027 is below the <0.11 BP4 threshold.
The governing MSH6 VCEP specification assigns BP4 Supporting to a missense variant with HCI prior probability <0.11.The exact HCI-PRIORS-MSH6 row for c.187T>C / p.S63P reports HCI prior probability 0.0027; the same row's custom PP2 score is 0.015 and is not the HCI prior probability used for this rule.The source registry reports a REVEL score of 0.276, but the governing VCEP HCI lookup directly pre-assigns BP4 Supporting and takes precedence over generic predictor thresholds.
Assessed · not applied · 3 not met · 11 not assessed
Pathogenic
PS1 Not assessed: no verified alternate-nucleotide p.Ser63Pro comparator was identified, and the same-residue search reported zero candidates with an HTTP 429 retrieval error.
PS2 Not assessed: no variant-specific proband, parental-confirmation, tumor, or de novo-point evidence is available for the PS2 thresholds.
PS3 Not assessed: no variant-specific functional assay result or calibrated functional odds are available for c.187T>C (p.Ser63Pro).
PM2 Not met: gnomAD v4.1 total allele frequency is 0.0000344201, exceeding the MSH6 PM2 threshold of 0.00002.
PM3 Not assessed: no affected-proband observation, pathogenic MSH6 allele, or confirmed in-trans phase is documented, so PM3 points cannot be assigned.
PM5 Not assessed: the p.Ser63Pro comparator search found zero same-residue candidates and an HTTP 429 error, so the required VCEP comparator and PP3 prerequisite remain unverified.
PP1 Not assessed: no variant-specific pedigree, meioses, relative genotypes, or Bayes likelihood ratio is available for the PP1 thresholds.
PP3 Not met: MSH6 HCI prior probability 0.0027 is below the >0.68 PP3-supporting threshold.
PP4 Not assessed: no variant-specific MSI-H tumor count or MSH6-consistent IHC result is available to compare with the VCEP's PP4 thresholds.
Benign
BA1 Not met: gnomAD v4.1 grpmax filtering allele frequency is 0.00055895, below the MSH6 BA1 threshold of 0.0022.
BS2 Not assessed: gnomAD reports zero homozygotes, but no qualifying in-trans co-occurrence, cancer age, CMMRD, or phase data are available.
BS3 Not assessed: no variant-specific proficient-function assay result or calibrated functional odds <=0.05 are available for c.187T>C (p.Ser63Pro).
BS4 Not assessed: no variant-specific non-segregation pedigree data or combined Bayes likelihood ratio is available for the BS4 thresholds.
BP5 Not assessed: no variant-specific MSS, preserved/discordant MMR IHC, BRAF V600E, or MLH1 methylation findings are documented for the BP5 thresholds.
N/A · 12 PVS1 · PS4 · PM1 · PM4 · PM6 · PP2 · PP5 · BP1 · BP2 · BP3 · BP6 · BP7
Research & evidence
Population frequency · supports benign
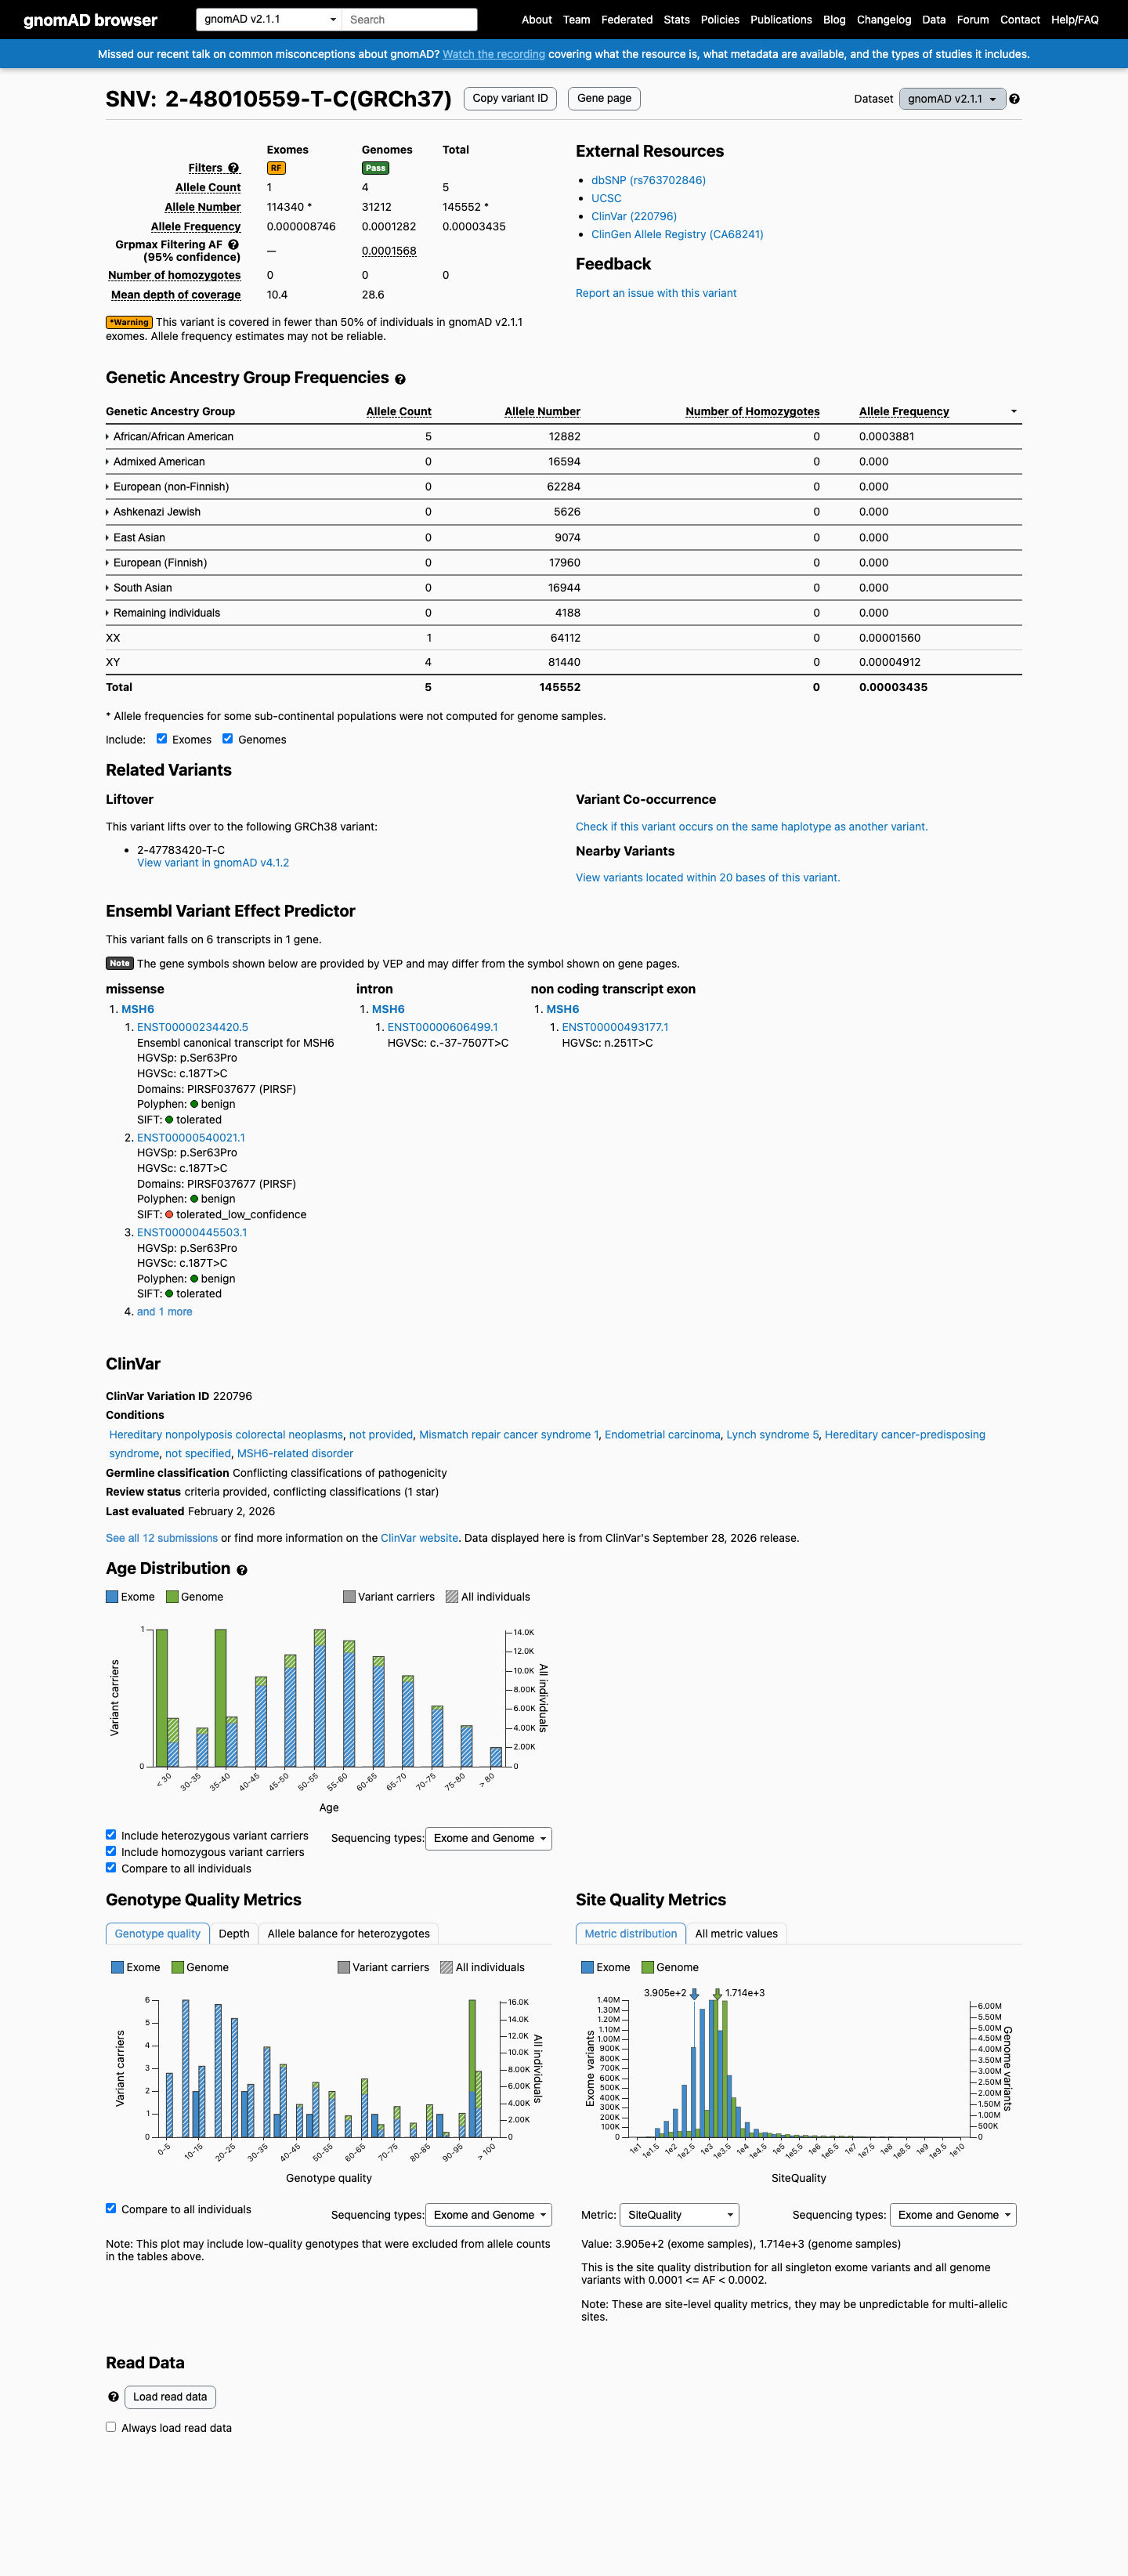
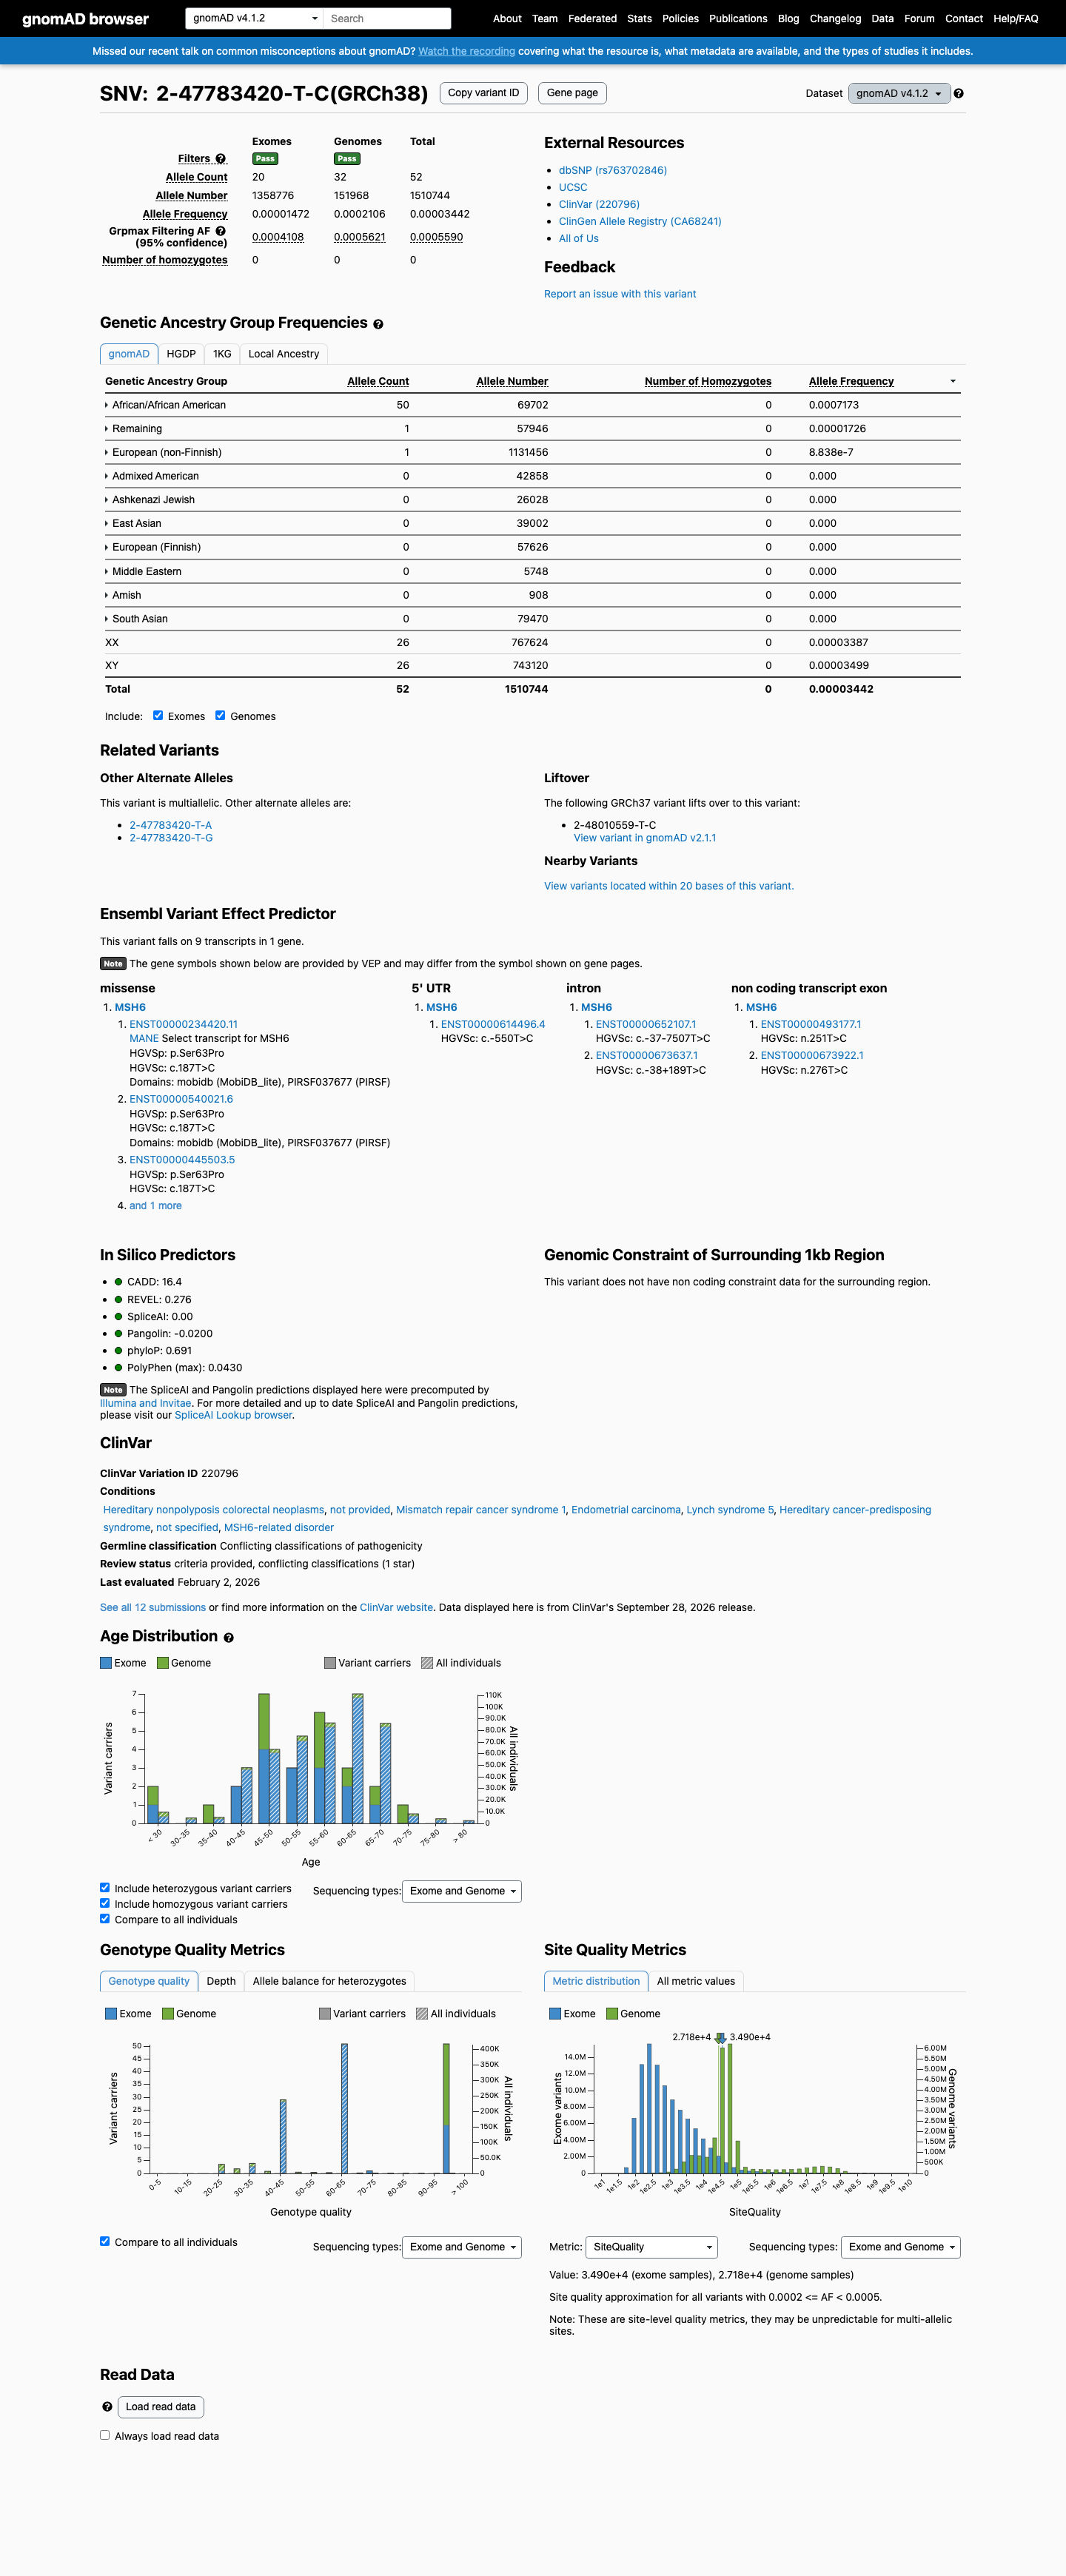
gnomAD v4.1 screenshot
gnomAD v4.1
gnomAD v2.1 screenshot
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 3.44201e-05; MAF= 0.00344%, 52/1510744 alleles, homozygotes = 0) and has highest observed frequency in the African/African American population (AF= 0.00071734; MAF= 0.07173%, 50/69702 alleles, homozygotes = 0); grpmax FAF= 0.00055895.
v2.1
This variant is present in gnomAD v2.1 (AF= 3.4352e-05; MAF= 0.00344%, 5/145552 alleles, homozygotes = 0) and has highest observed frequency in the African/African American population (AF= 0.000388138; MAF= 0.03881%, 5/12882 alleles, homozygotes = 0); grpmax FAF= 0.00015682.
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.0034% · 52 / 1,510,744
0 hom · FAF 0.056%
African/African American
50 / 69,702
0.072%
Remaining individuals
1 / 57,946
0.0017%
European (non-Finnish)
1 / 1,131,456
8.8e-05%
+ 7 not observed (Admixed American, European (Finnish), Amish, East Asian, Middle Eastern, South Asian, Ashkenazi Jewish)
gnomAD v2.1
0.0034% · 5 / 145,552
0 hom · FAF 0.016%
African/African American
5 / 12,882
0.039%
+ 7 not observed (Admixed American, Ashkenazi Jewish, East Asian, European (Finnish), European (non-Finnish), Remaining individuals, South Asian)
gnomAD Canada 🇨🇦
Absent · 0 / ?
0 hom
Not observed in any ancestry group.
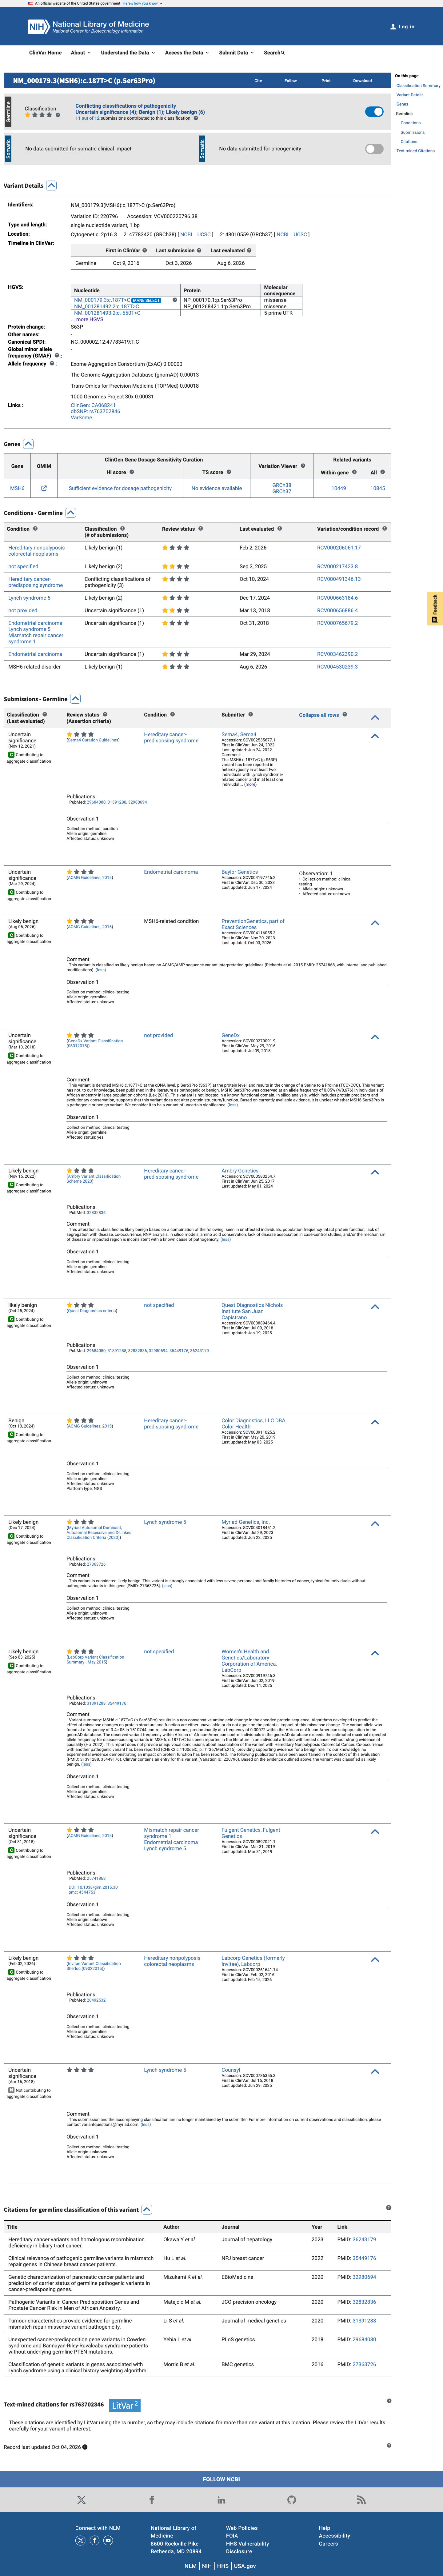
ClinVar screenshot
ClinVar
This variant has been reported in ClinVar as Likely benign (5 clinical laboratories) and as Uncertain significance (4 clinical laboratories) and as likely benign (1 clinical laboratory) and as Benign (1 clinical laboratory). (ClinVarID = 220796)
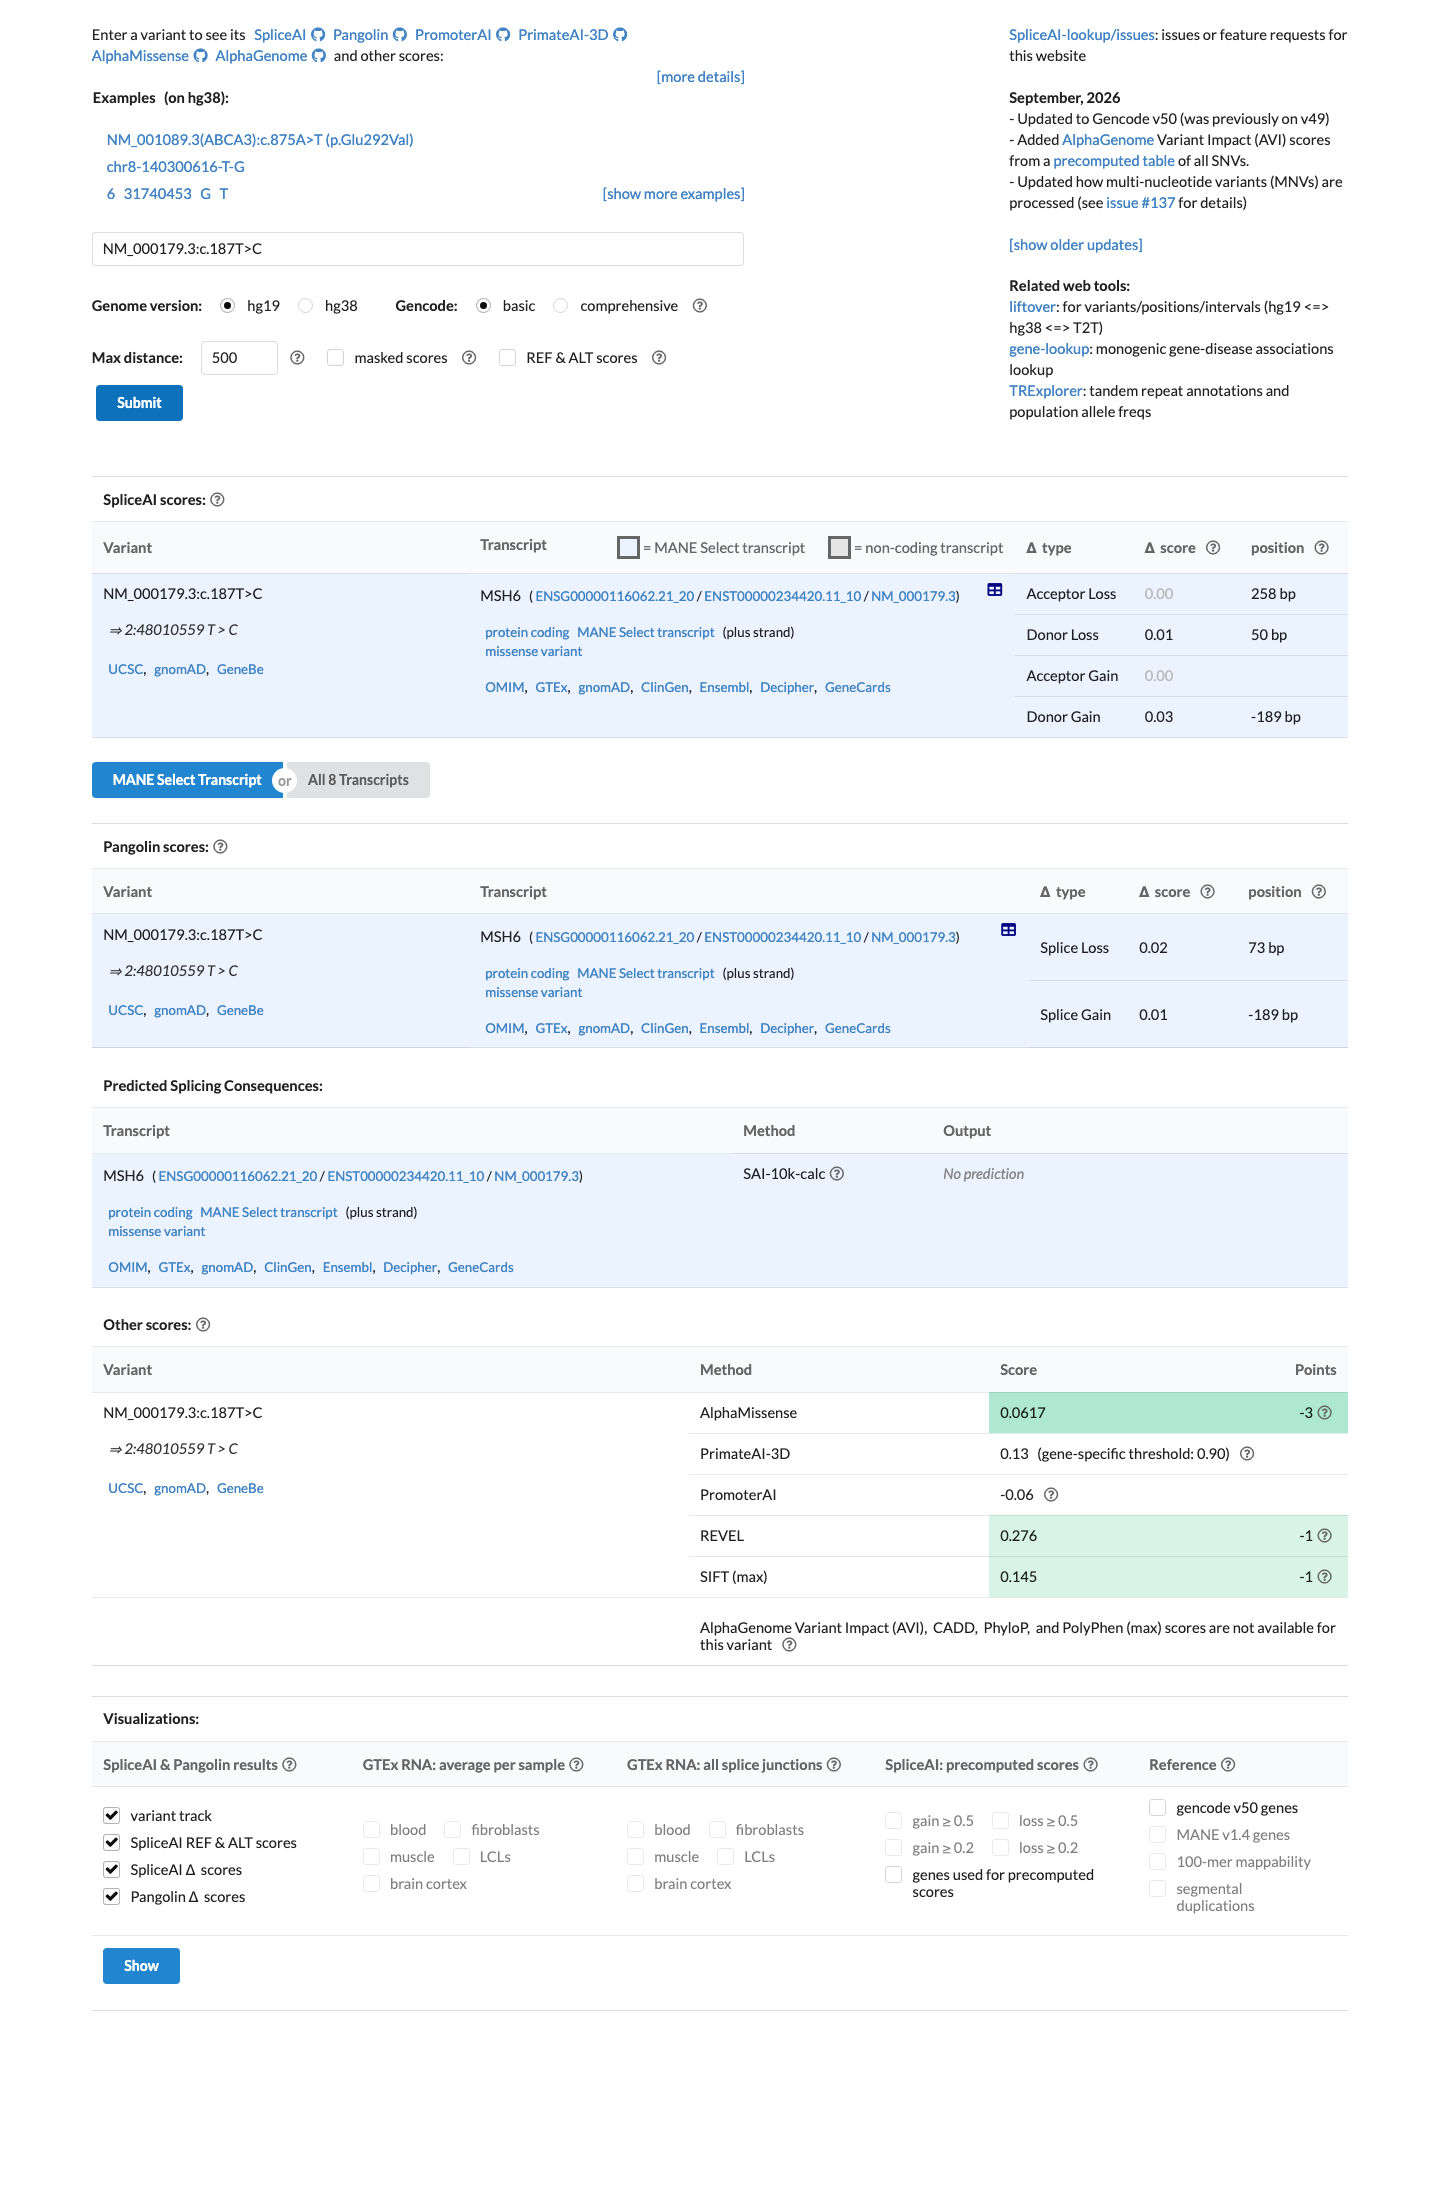
SpliceAI screenshot
In silico
SpliceAI predicts no significant splice impact for this variant (max delta score = 0.03). REVEL score = 0.276. BayesDel score = -0.26367. HCI prior probability for pathogenicity = 0.0027. Custom PP2 score = 0.015.
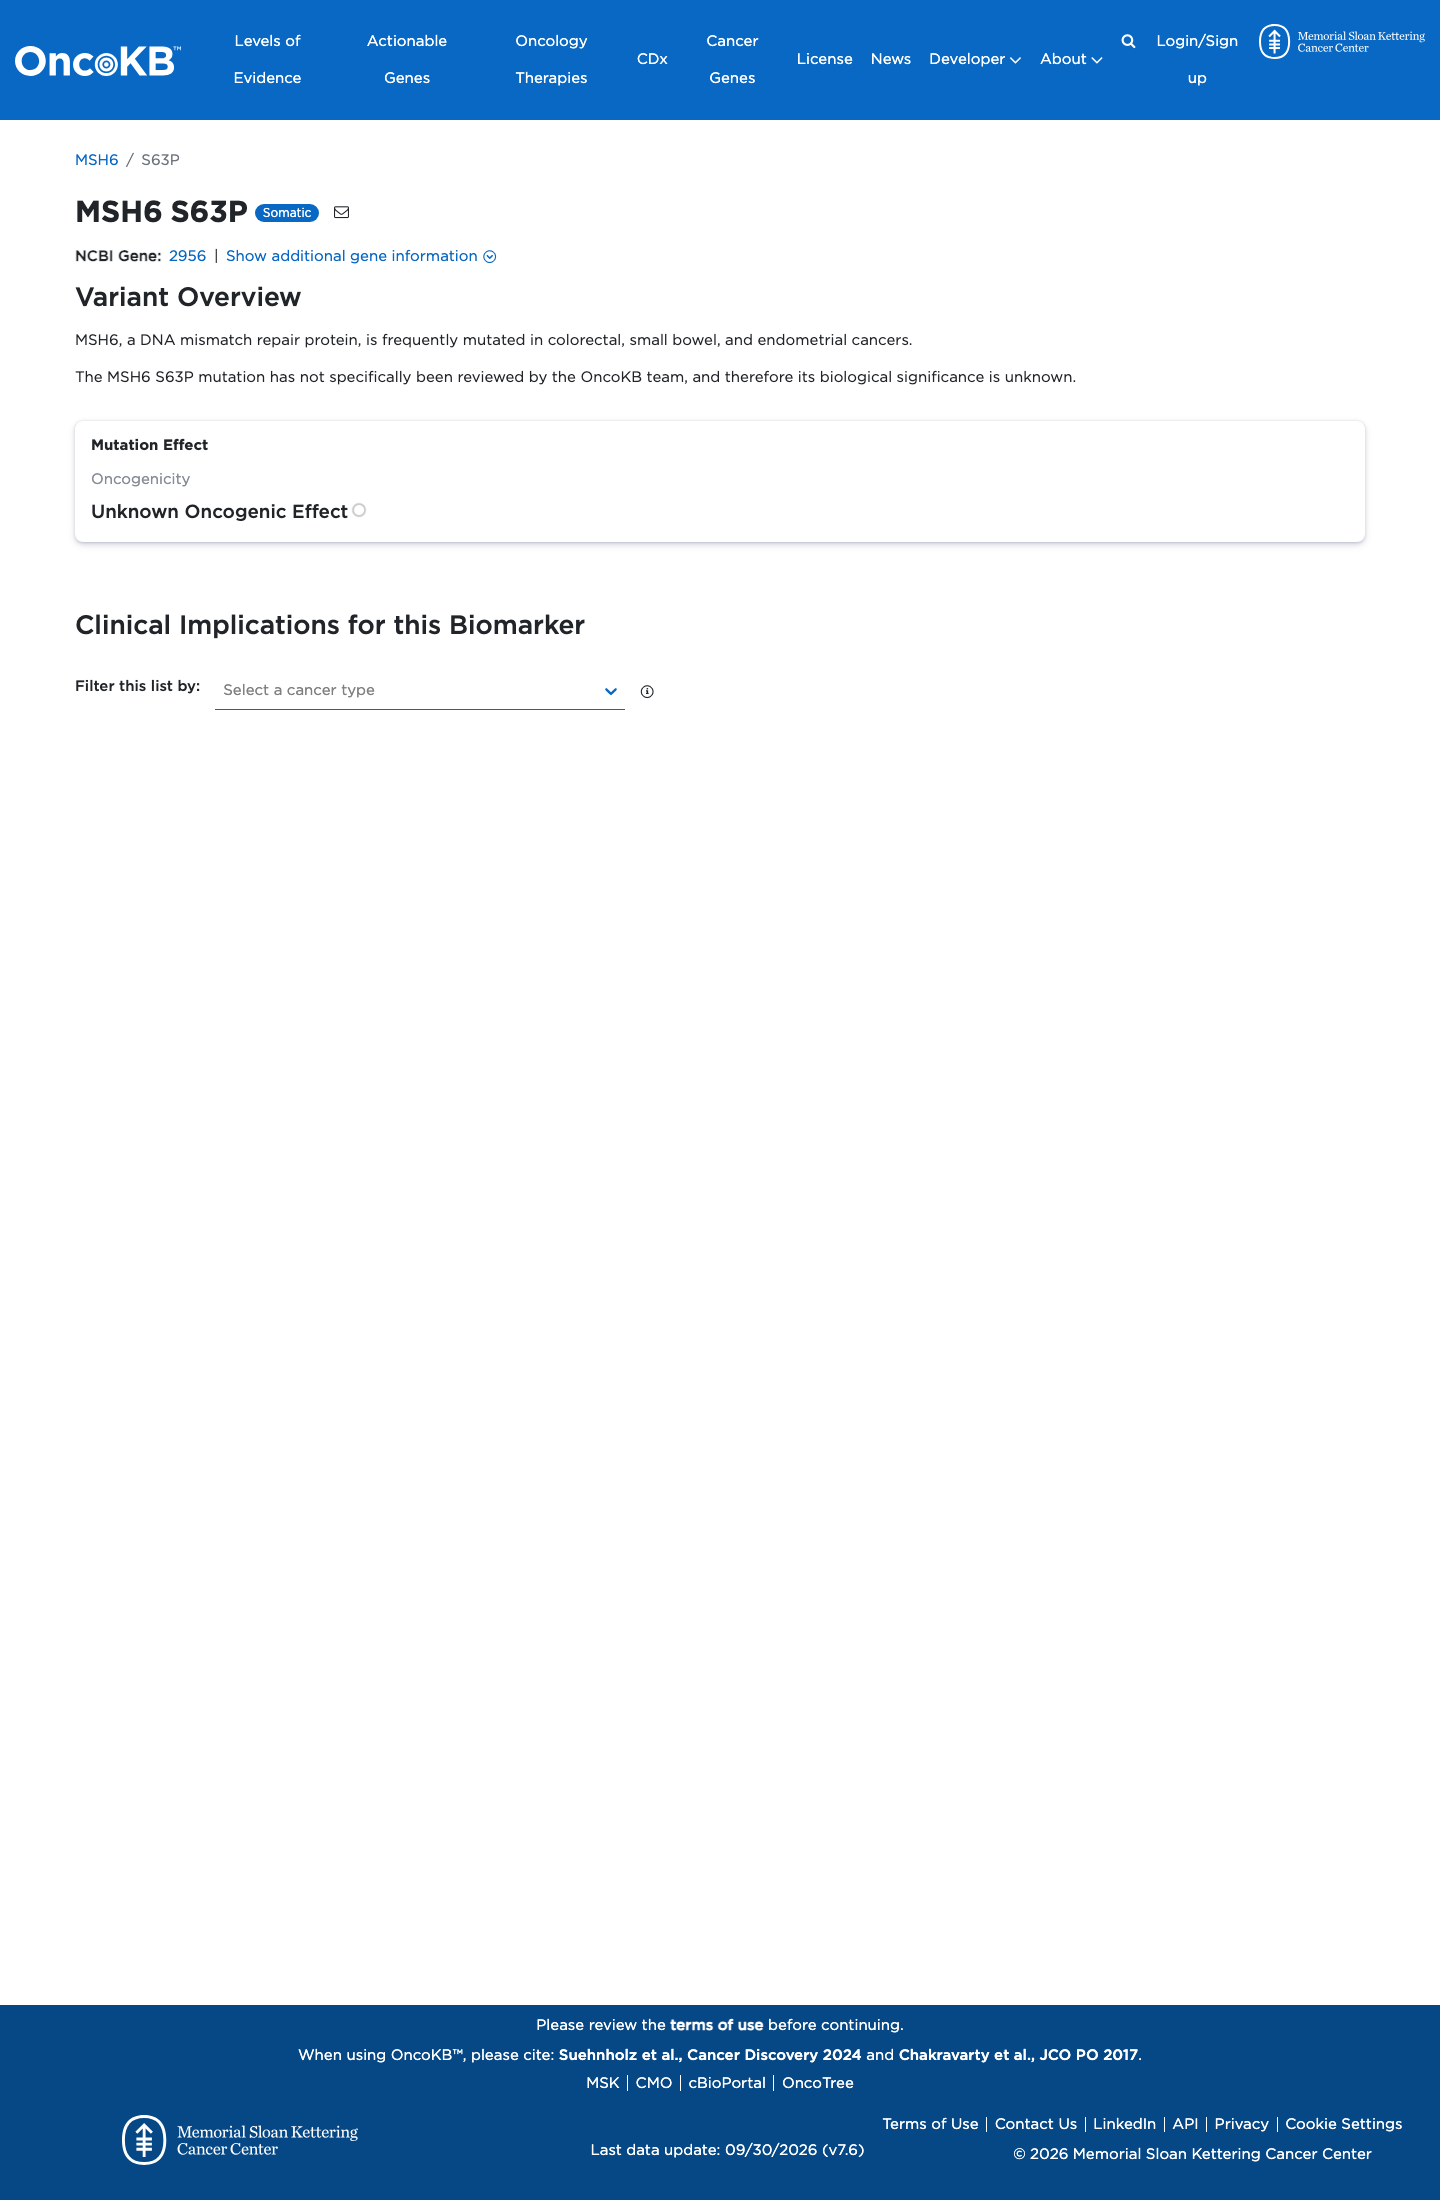
Functional / OncoKB screenshot
Functional Unknown Oncogenic Effect
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. MSH6, a DNA mismatch repair protein, is frequently mutated in colorectal, small bowel, and endometrial cancers.
OncoKB ↗
COSMIC screenshot
COSMIC
Cancer hotspots screenshot
Cancer hotspots
Somatic evidence Not in COSMIC / hotspots
COSMIC
This variant does not lie in a statistically significant hotspot. This variant has not previously been reported in somatic cancers (COSMIC).
Hotspots
This variant does not lie in a statistically significant hotspot.
Sources & reference links
9Sources
CSpec VCEP
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 8 PMIDs not cited in assessment
22167527 ↗ Identification of individuals at risk for Lynch syndrome using targeted evaluati CLINVAR
23788249 ↗ ACMG recommendations for reporting of incidental findings in clinical exome and genome sequencing. CLINVAR
25356965 ↗ ACMG policy statement: updated recommendations regarding analysis and reporting CLINVAR
25452455 ↗ Hereditary colorectal cancer syndromes: American Society of Clinical Oncology Cl CLINVAR
25741868 ↗ Standards and guidelines for the interpretation of sequence variants: a joint co CLINVAR
26467025 ↗ A Standardized DNA Variant Scoring System for Pathogenicity Assessments in Mende CLINVAR
27363726 ↗ Classification of genetic variants in genes associated with Lynch syndrome using a clinical history weighting algorithm. CLINVAR
27854360 ↗ Recommendations for reporting of secondary findings in clinical exome and genome CLINVAR