STK11 encodes a serine/threonine kinase that acts as a tumor suppressor, regulating cell polarity, energy metabolism, and stress responses through the AMPK signaling pathway. Germline mutations in this gene cause Peutz-Jeghers syndrome, an inherited condition marked by gastrointestinal polyps, mucocutaneous pigmentation, and an elevated risk of several cancers. Loss of STK11 function also occurs in cancers of the lung, pancreas, breast, cervix, liver, and other tissues, where it promotes tumor growth.
This variant
This STK11 missense variant occurs in a tumor-suppressor gene in which loss-of-function causes autosomal-dominant Peutz-Jeghers syndrome and contributes to tumor development through impaired AMPK-pathway regulation.
Transcript
NM_000455.5
HGVS · transcript:coding
NM_000455.5:c.911G>A
GRCh38
chr19:1221997 G>A
GRCh37
chr19:1221996 G>A
VUS: PM2 at supporting strength is the only applied criterion, which does not meet generic ACMG/AMP thresholds for a definitive pathogenic or benign classification.
Classification rationale
PM2VUS
STK11 c.911G>Amissense · exon 7
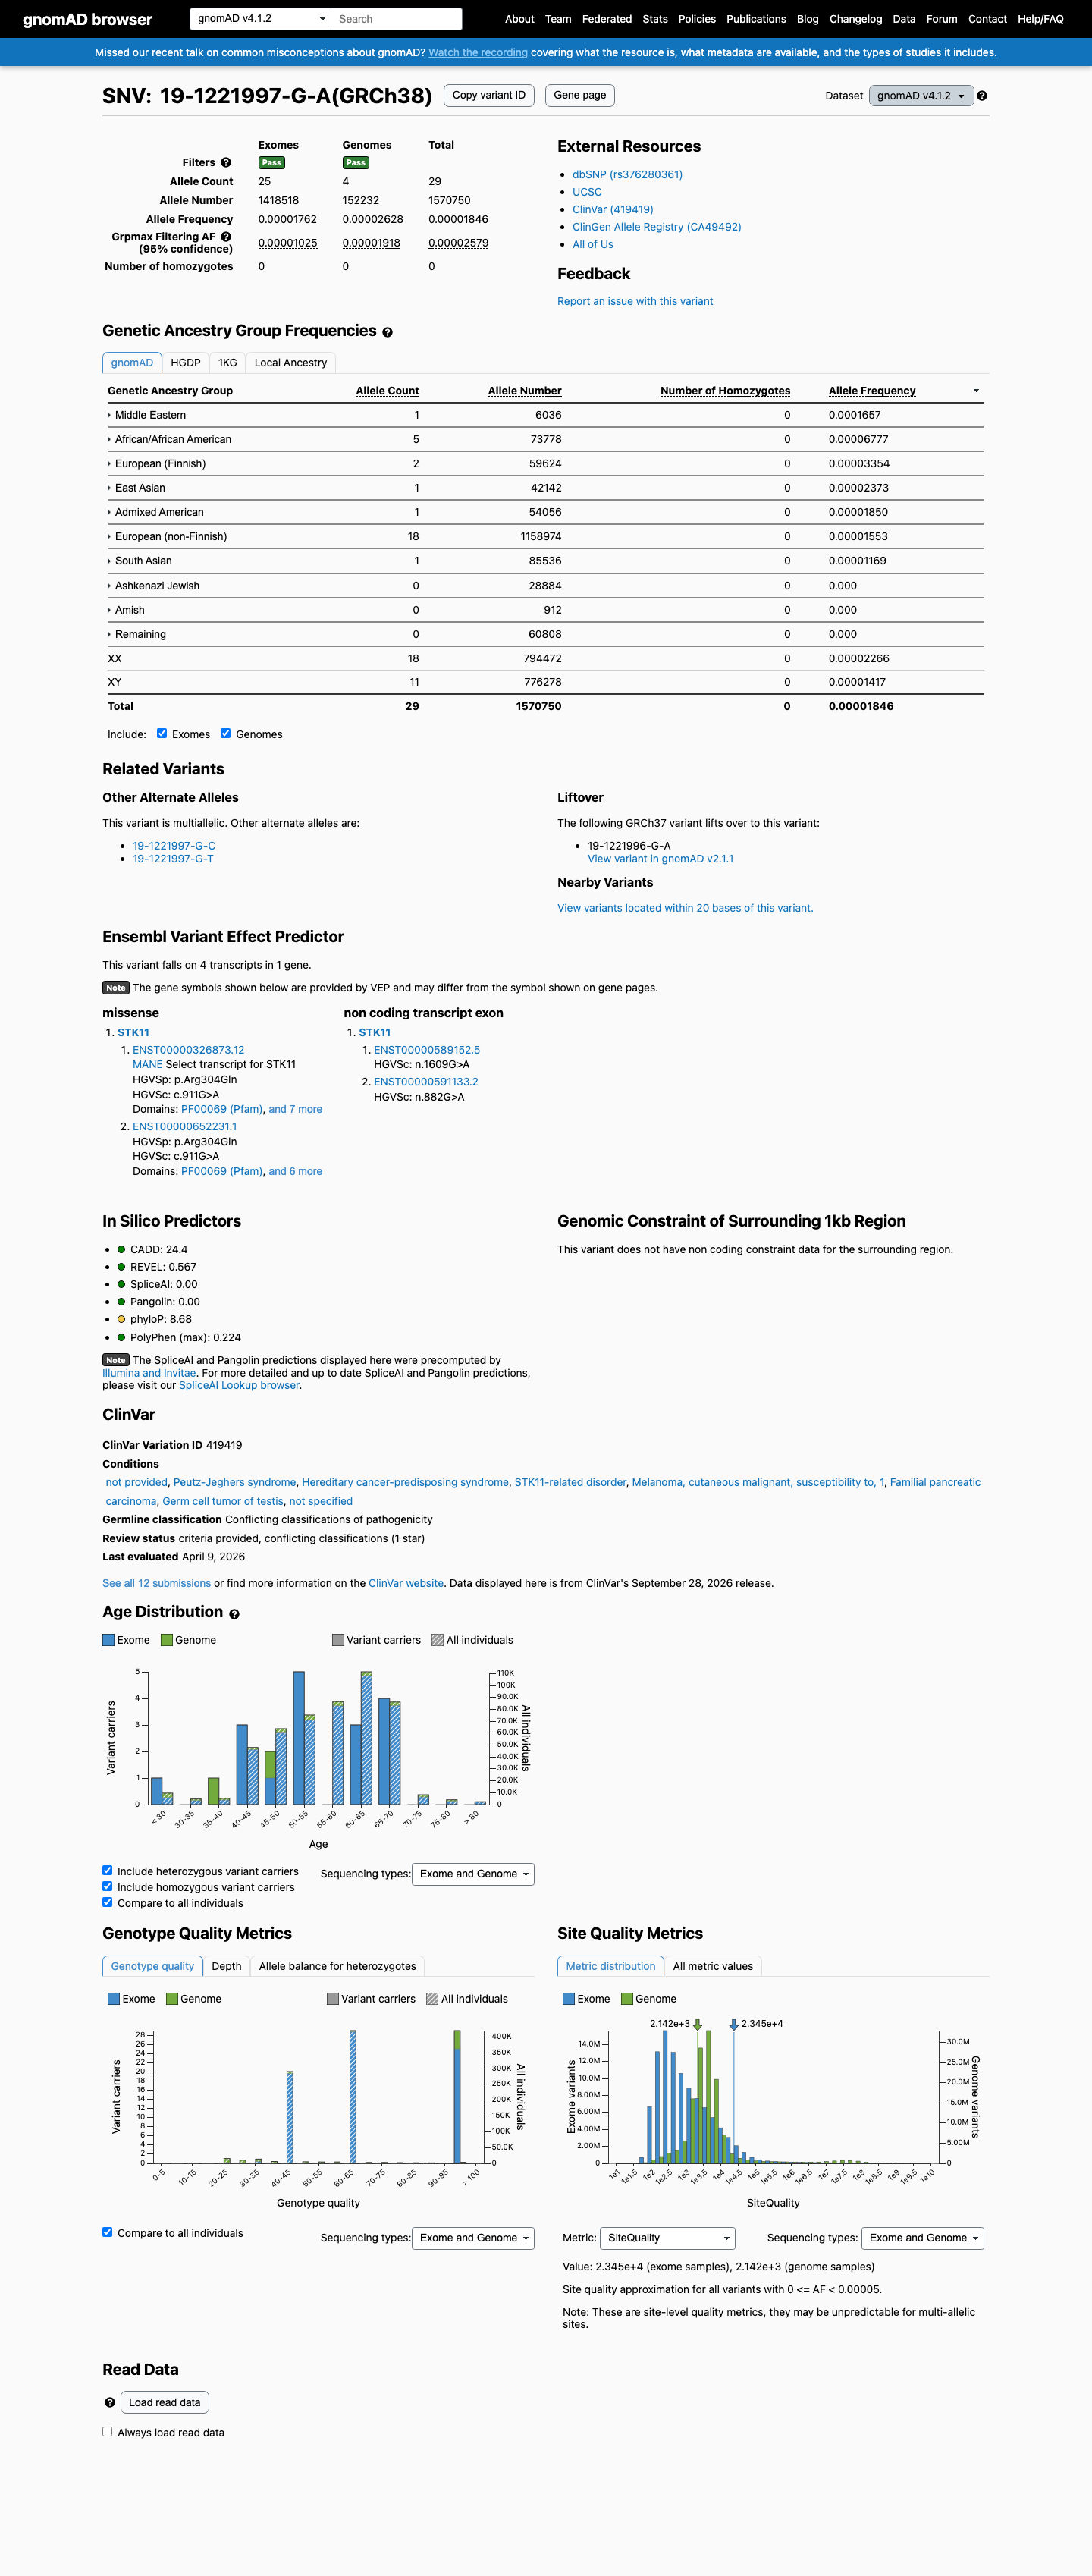
PM2 supporting: gnomAD v4.1 allele frequency is below 0.0001 with zero homozygotes.
PM2→VUS
LYFE Sciences is an AI system, and it can make mistakes. Criteria
may be applied incorrectly, sources may be misread, and a confident-looking
classification can still be wrong. Double-check every criterion and
its underlying evidence before relying on any call.
Gene diagram
· NM_000455.5 · variants mapped to exon structure
STK11NM_000455.5
Fetching transcript structure from UCSC…
Exons
—
Transcript span
—
Strand
—
Variants mapped
—
Source
UCSC ncbiRefSeqCurated
All variants in STK11—click a row to locate it on the plot · use the link column to open its page
Protein
Location
Classification
Link
Applied criteria · 1 applied · 22 assessed
MetEvidence satisfies this criterion.
Not metEvaluated against available evidence; threshold not reached.
Not assessedApplies in principle, but no evidence was found to evaluate it.
N/ADoesn't apply to this variant type.
Applied · 1
Strength Supporting Moderate Strong Very strong
✓
PM2supportingPathogenic
Met at supporting: gnomAD v4.1 all-comers AF 1.84625e-05 is below the generic PM2 threshold of 0.0001, with zero homozygotes.
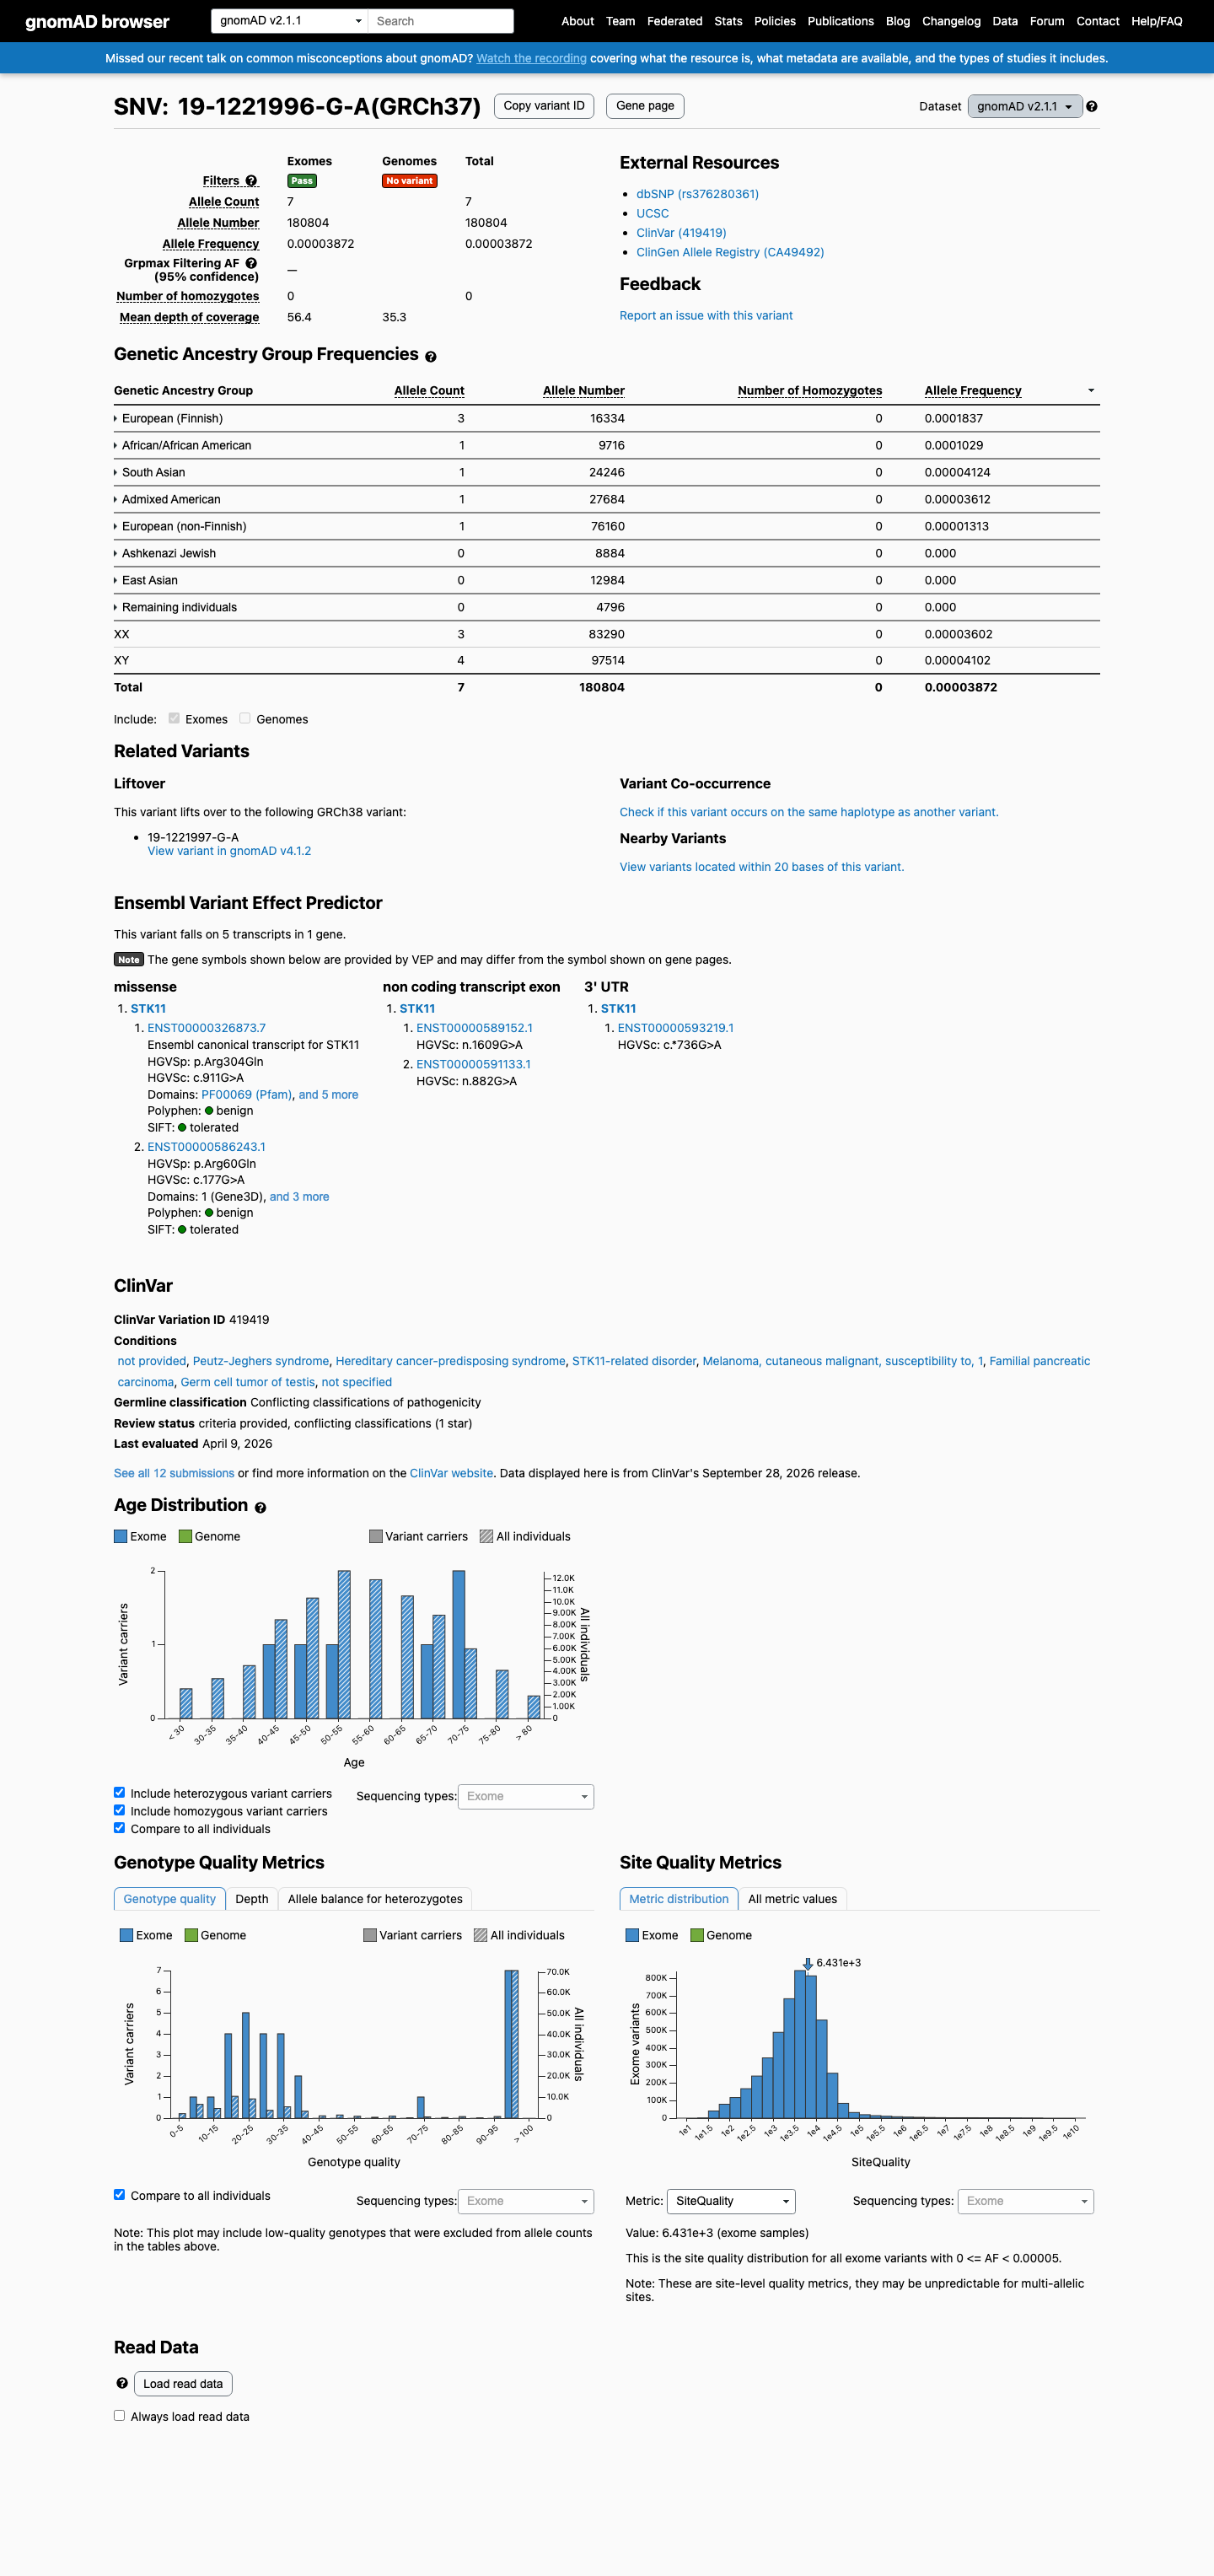
No applicable STK11-specific VCEP specification was identified; the supplied generic PM2 threshold is allele frequency <=0.0001 at supporting strength.Default all-comers source, gnomAD v2.1 exomes: AF 3.871595761155727e-05 (7/180804), 0 homozygotes.Default all-comers source, gnomAD v4.1 exomes plus genomes: AF 1.8462517905459174e-05 (29/1570750), 0 homozygotes, and group maximum FAF 2.579e-05.
Assessed · not applied
· 7 not met · 15 not assessed
Pathogenic
PS1PS1 could not be assessed because its agent did not produce valid output.
PS2Not assessed: no parental testing or confirmed de novo observation is documented for the present c.911G>A (p.Arg304Gln) variant.
PS3Not assessed: no validated functional assay tested the exact R304Q variant, while available assays studied different STK11 substitutions such as R304W.
PS4Not assessed: no exact-variant case-control counts, enrichment statistic, odds ratio, or affected-case versus control comparison is available.
PM1PM1 could not be assessed because its agent did not produce valid output.
PM5PM5 could not be assessed because its agent did not produce valid output.
PM6Not assessed: no apparently de novo occurrence is reported for c.911G>A (p.Arg304Gln), and parental testing is unavailable.
PP1Not assessed: zero informative meioses or affected relatives carrying c.911G>A (p.Arg304Gln) are documented.
PP2PP2 could not be assessed because its agent did not produce valid output.
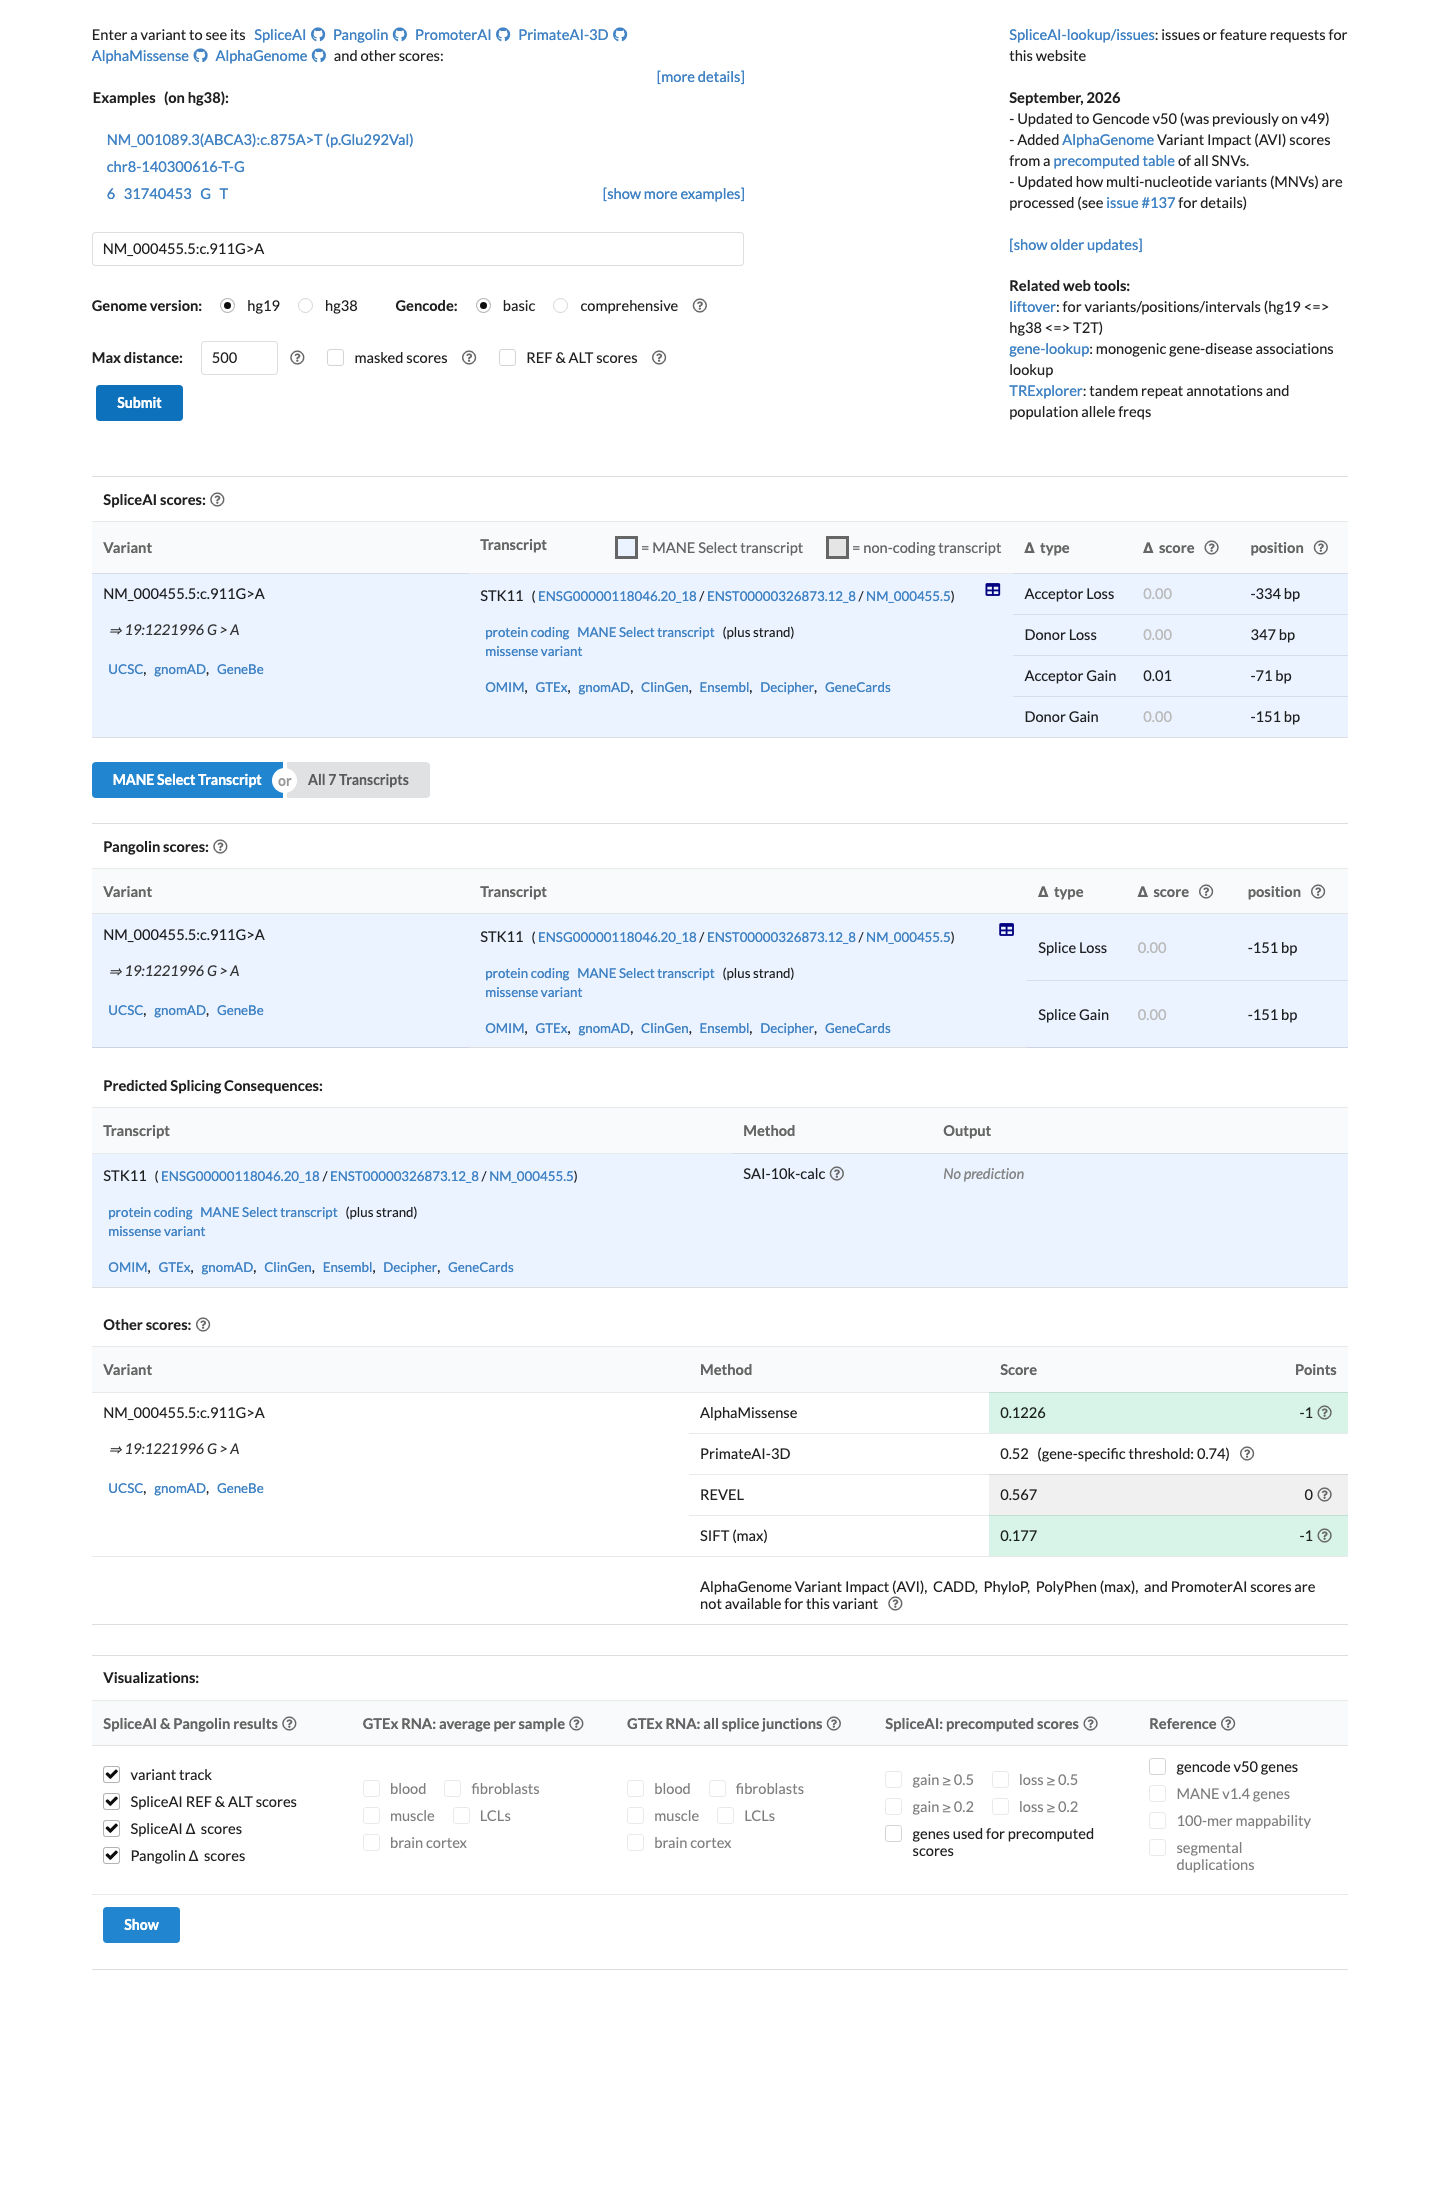
PP3Not met: REVEL 0.567 is below the supporting PP3 threshold of 0.644 for missense variants.
PP4Not assessed: no patient phenotype or phenotype-specific diagnostic findings are documented for comparison with an STK11-related disorder.
PP5Not met: the exact ClinVar record has zero expert-panel submissions and no expert-panel Pathogenic or Likely pathogenic classification.
Benign
BA1Not met: all-comers gnomAD v4.1 AF 1.84625e-05 is far below the generic BA1 threshold of 0.05.
BS1Not met: the highest observed gnomAD ancestry-specific AF, 0.000183666, remains below the generic BS1 threshold of 0.01.
BS2Not met: gnomAD v2.1 and v4.1 each report 0 homozygotes, so no healthy-homozygote evidence supports BS2.
BS3Not assessed: no validated benign functional assay tested exact STK11 R304Q with evidence of preserved biologically relevant function.
BS4Not assessed: no tested unaffected relatives or phenotype-informed non-segregation observations are documented for c.911G>A (p.Arg304Gln).
BP1BP1 could not be assessed because its agent did not produce valid output.
BP2Not assessed: no documented cis/trans phase or co-occurrence observation with a pathogenic STK11 variant is available.
BP4Not met: REVEL 0.567 exceeds the supporting BP4 threshold of 0.29 for missense variants.
BP5Not assessed: no affected individual with a documented alternate molecular diagnosis is available to support BP5.
BP6Not met: one ordinary laboratory submission is Likely benign, but the exact ClinVar record has zero expert-panel submissions.
N/A · 5PVS1 · PM3 · PM4 · BP3 · BP7
Research & evidence
Population frequency
gnomAD v4.1
gnomAD v2.1
v4.1
This variant is present in gnomAD v4.1 (AF= 1.84625e-05; MAF= 0.00185%, 29/1570750 alleles, homozygotes = 0) and has highest observed frequency in the Middle Eastern population (AF= 0.000165673; MAF= 0.01657%, 1/6036 alleles, homozygotes = 0); grpmax FAF= 2.579e-05.
v2.1
This variant is present in gnomAD v2.1 (AF= 3.8716e-05; MAF= 0.00387%, 7/180804 alleles, homozygotes = 0) and has highest observed frequency in the European (Finnish) population (AF= 0.000183666; MAF= 0.01837%, 3/16334 alleles, homozygotes = 0).
🇨🇦 CA
Not available in gnomAD-Canada v1.0.
Allele frequency by ancestry
three datasets · side by side
gnomAD v4.1
0.0018%
· 29 / 1,570,750
0 hom · FAF 0.0026%
Middle Eastern
1 / 6,036
0.017%
African/African American
5 / 73,778
0.0068%
European (Finnish)
2 / 59,624
0.0034%
East Asian
1 / 42,142
0.0024%
Admixed American
1 / 54,056
0.0018%
European (non-Finnish)
18 / 1,158,974
0.0016%
South Asian
1 / 85,536
0.0012%
+ 3 not observed (Remaining individuals, Amish, Ashkenazi Jewish)
gnomAD v2.1
0.0039%
· 7 / 180,804
0 hom
European (Finnish)
3 / 16,334
0.018%
African/African American
1 / 9,716
0.01%
South Asian
1 / 24,246
0.0041%
Admixed American
1 / 27,684
0.0036%
European (non-Finnish)
1 / 76,160
0.0013%
+ 3 not observed (Ashkenazi Jewish, East Asian, Remaining individuals)
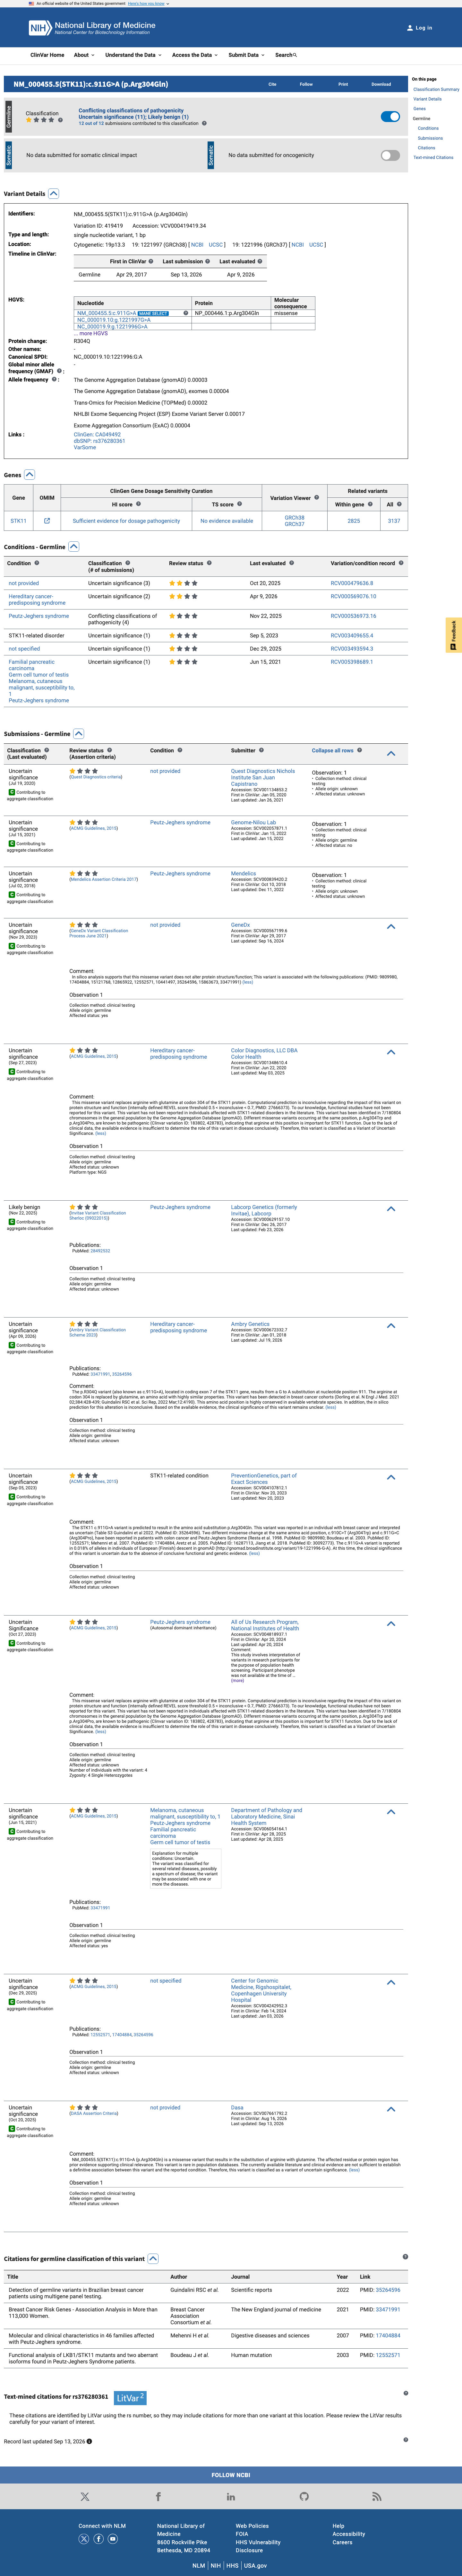
This variant has been reported in ClinVar as Uncertain significance (10 clinical laboratories) and as Likely benign (1 clinical laboratory) and as Uncertain Significance (1 clinical laboratory). (ClinVarID = 419419)
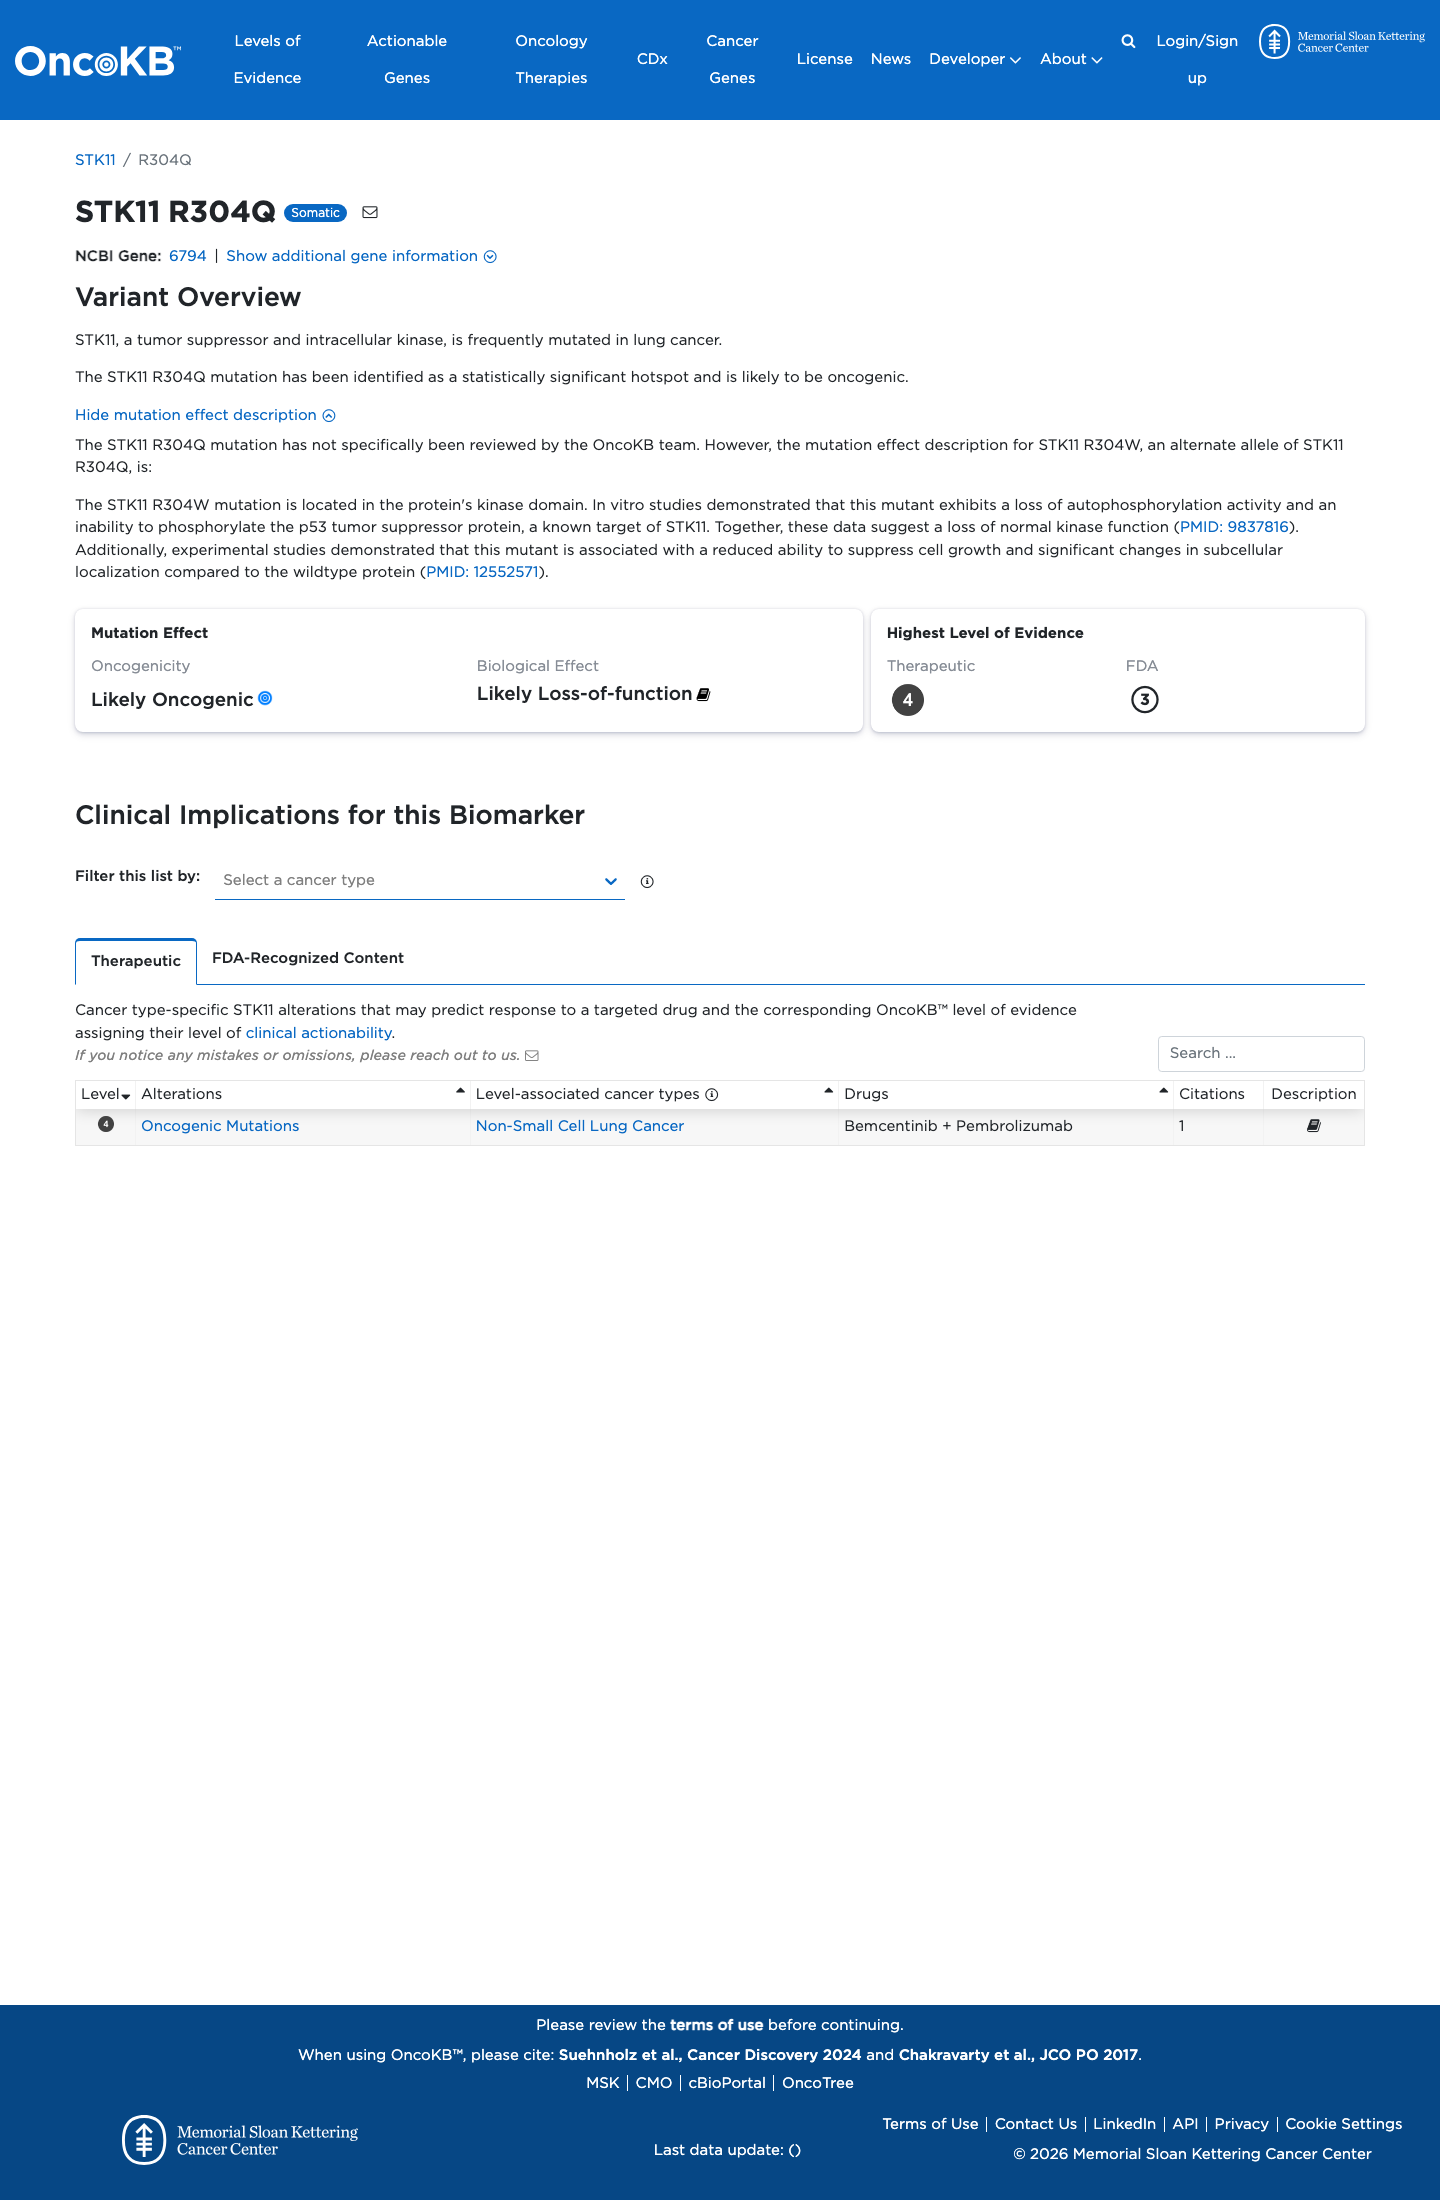
OncoKB did not identify variant-specific reviewed functional evidence for this variant; gene-level curated context is available for reviewer follow-up. STK11, a tumor suppressor and intracellular kinase, is frequently mutated in lung cancer.
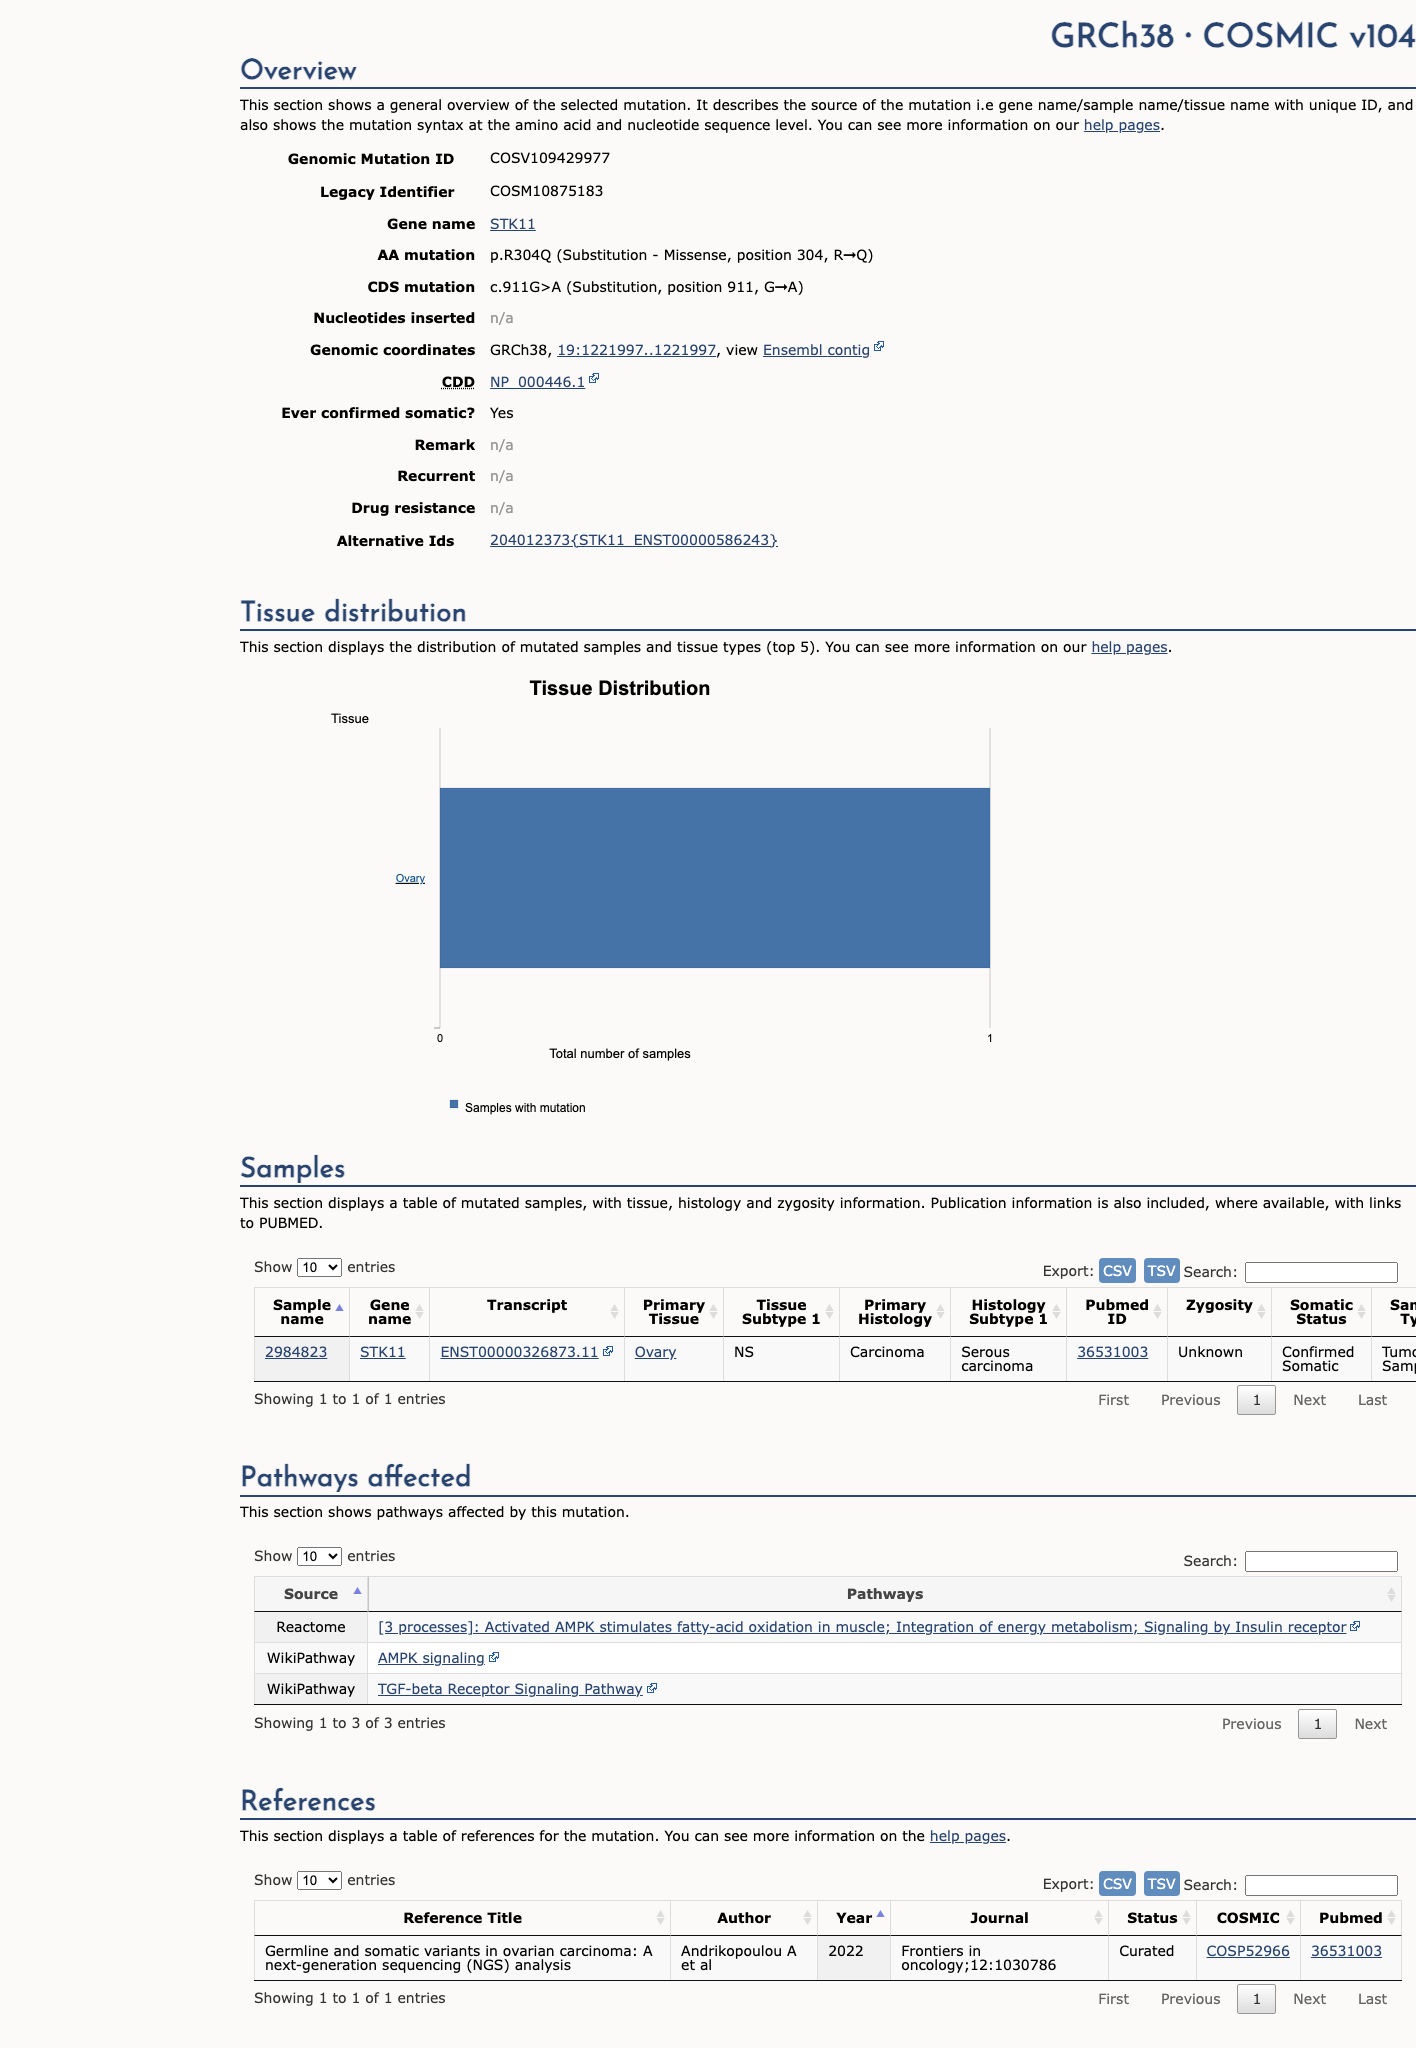
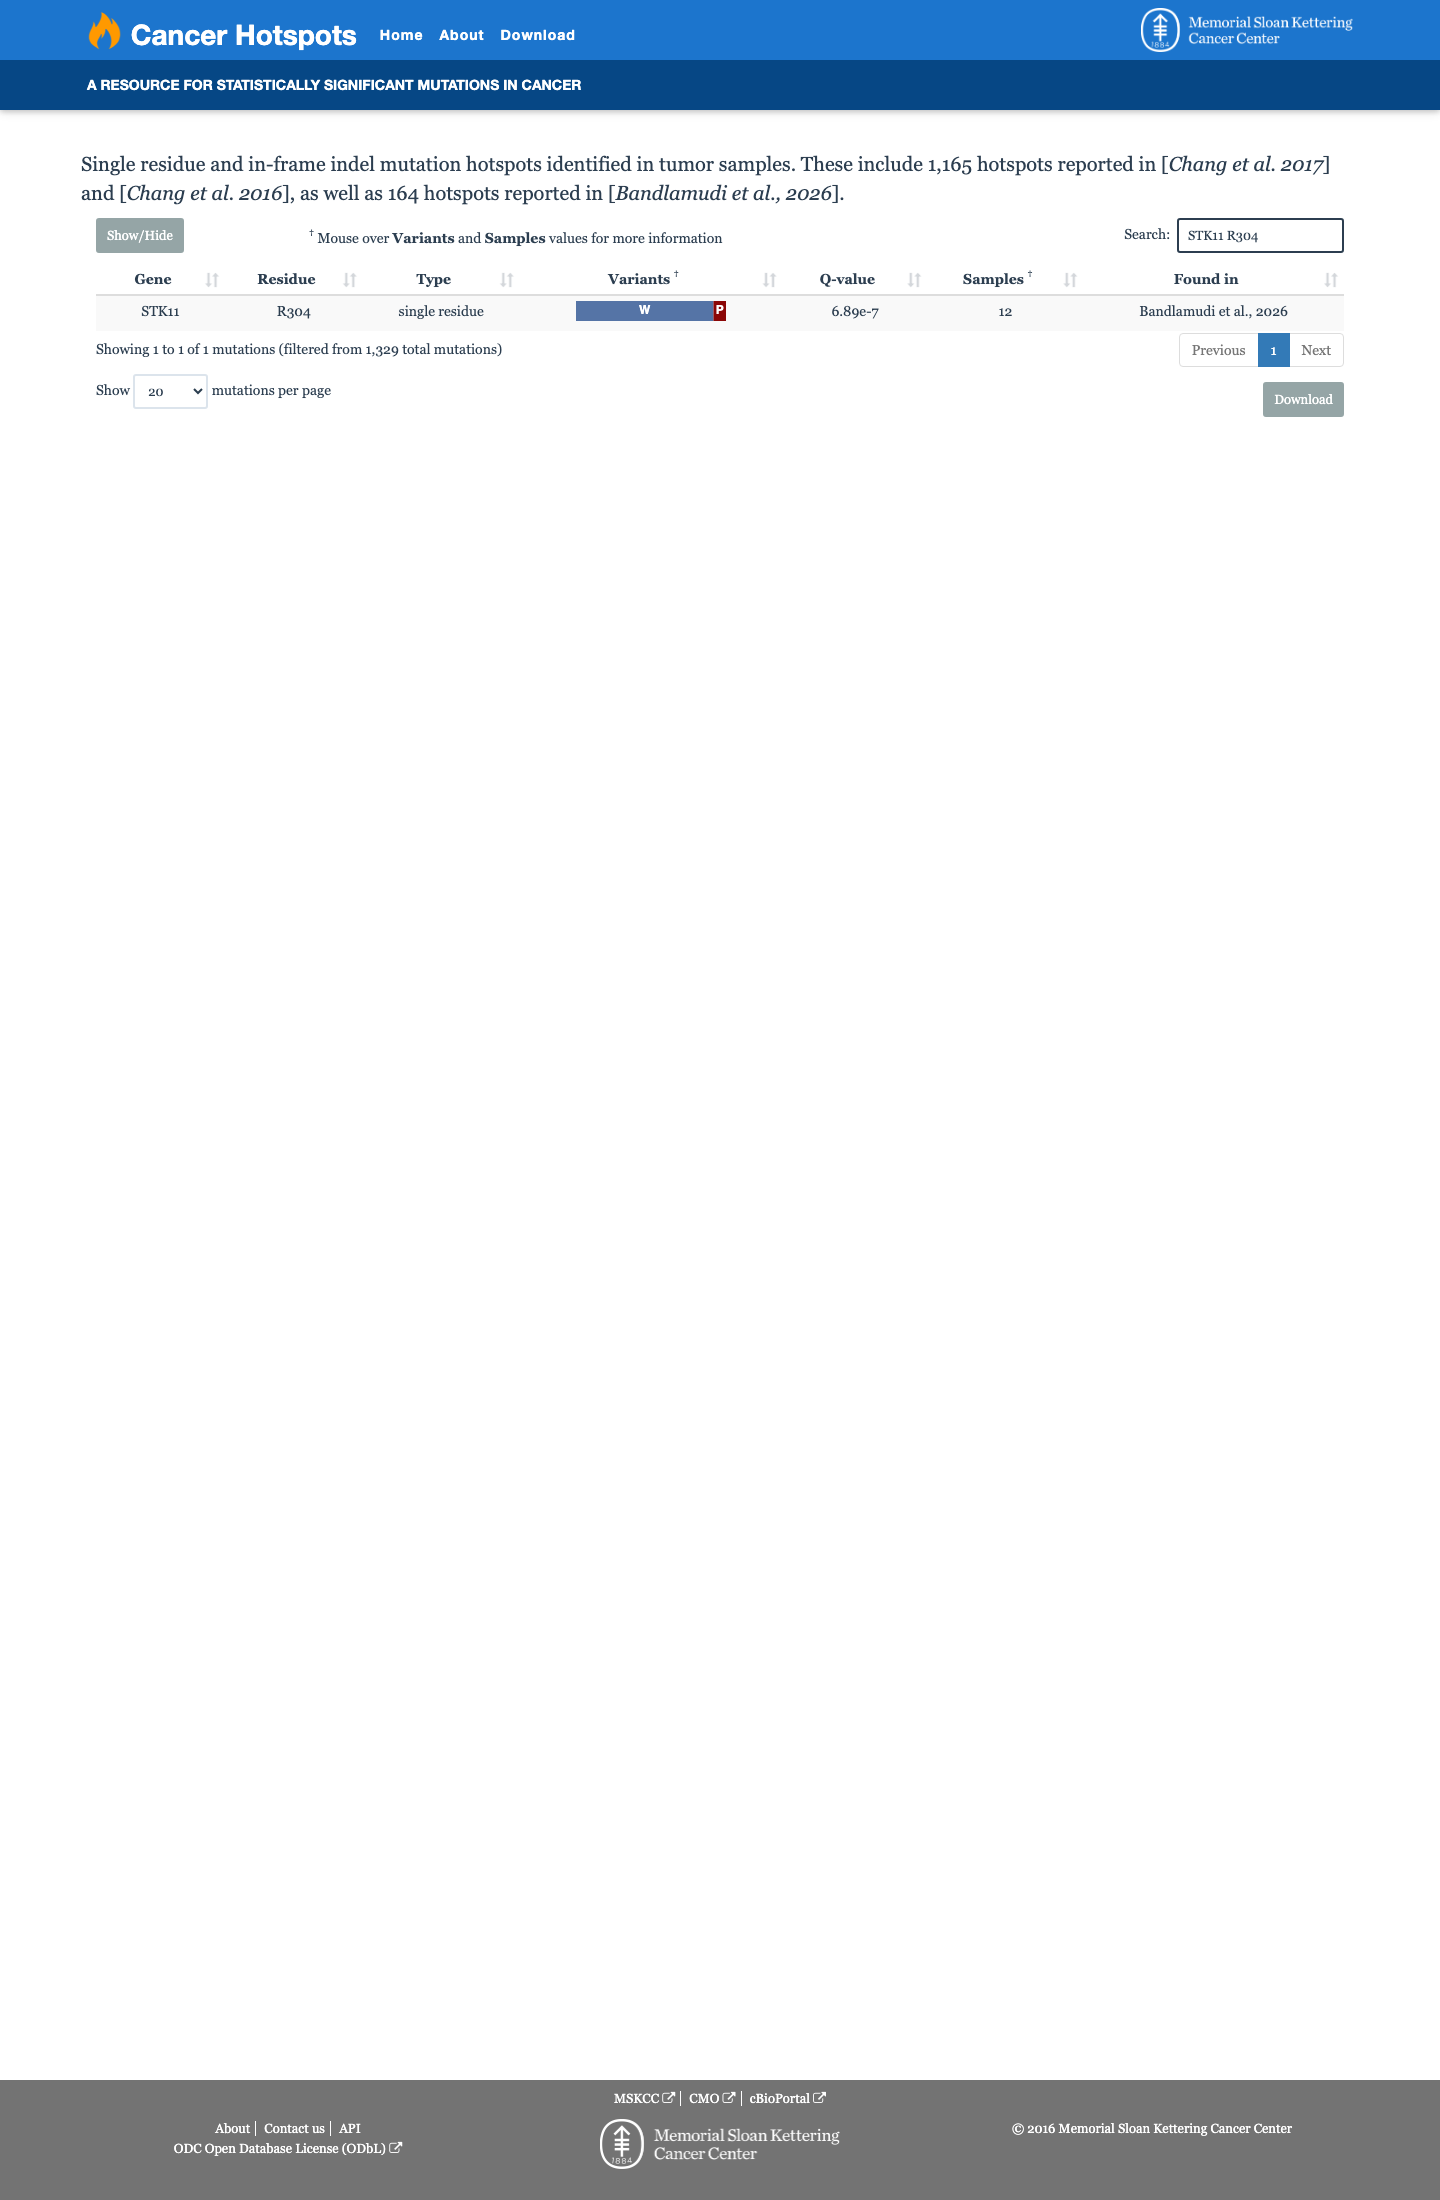
This variant lies in a statistically significant hotspot. This variant has previously been reported in somatic cancers (COSMIC; COSV109429977, n = 1 times).
Hotspots
This variant lies in a statistically significant hotspot.
Each card is an audit: what was searched, what was found, whether it names the variant, which criteria it fed, and why. 7 further PMIDs triaged but not cited — see Sources & references.
Rule & framework references · cited for criterion definitions, not variant evidence
25741868 ↗Standards and guidelines for the interpretation of sequence variants: a joint consensus recommendation of the American College of Medical Genetics and Genomics and the Association for Molecular Pathology.
Sources & reference links
8Sources
ClinVar
gnomAD v2.1
gnomAD v4.1
gnomAD-Canada
SpliceAI
OncoKB
COSMIC
Cancer hotspots
Triaged references · 7 PMIDs not cited in assessment
12552571 ↗Functional analysis of LKB1/STK11 mutants and two aberrant isoforms found in Peutz-Jeghers Syndrome patients.ONCOKB
9837816 ↗Loss of LKB1 kinase activity in Peutz-Jeghers syndrome, and evidence for allelic and locus heterogeneity.ONCOKB
17404884 ↗Molecular and clinical characteristics in 46 families affected with Peutz-Jeghers syndrome.CLINVAR
26467025 ↗A Standardized DNA Variant Scoring System for Pathogenicity Assessments in Mendelian Disorders.CLINVAR
23788249 ↗ACMG recommendations for reporting of incidental findings in clinical exome and genome sequencing.CLINVAR
25356965 ↗ACMG policy statement: updated recommendations regarding analysis and reporting of secondary findings in clinical genome-scale sequencing.CLINVAR
28492532 ↗Sherloc: a comprehensive refinement of the ACMG-AMP variant classification criteria.CLINVAR